Home » Uncategorized (Page 80)
Category Archives: Uncategorized
 DRUG APPROVALS BY DR ANTHONY MELVIN CRASTO
.....FOR BLOG HOME CLICK HERE
Flag and hits
ORGANIC SPECTROSCOPY

SUBSCRIBE

 Subscribe in a reader
Enter your email address:
Delivered by FeedBurner
Subscribe to New Drug Approvals by Email


Cymipristone
Cymipristone
- Estra-4,9-dien-3-one, 11-[4-(cyclohexylmethylamino)phenyl]-17-hydroxy-17-(1-propynyl)-, (11β,17β)- (9CI)
- (11β,17β)-11-[4-(Cyclohexylmethylamino)phenyl]-17-hydroxy-17-(1-propyn-1-yl)estra-4,9-dien-3-one
- Saimisitong
NDA Filed china
Shanghai Siniwest Pharmaceutical Chemical Technology Co., Ltd., Shanghai Zhongxi Pharmaceutical Co. Ltd., Xianju Pharmaceutical Co., Ltd,
A progesterone receptor antagonist potentially for termination of intrauterine pregnancy.

CAS No.329971-40-6
- Molecular FormulaC34H43NO2
- Average mass497.711 Da
- Steroid Compounds, a Method for Preparation thereof, Pharmaceutical Compositions Containing the Same and Use thereof
-
This invention relates to steroid compounds and pharmaceutical acceptable salts thereof, a method for preparation thereof, pharmaceutical compositions containing the same as active component, and their use in the preparation of medicines for treating diseases associated with progestogen dependence and for fertility control, abortion or contraception and for anticancer use.
-
Mifepristone (11β-[4-(N,N-dimethylamino)phenyl]-17α-(1-propinyl)-17β-hydroxy-4,9-estradiene-3-one) is a steroid compound which is disclosed in French Patent No. 2,497,807 to Rousell-Uclaf, published May 31, 1983. It is the first progesterone receptor antagonist put into clinical application and is a new type of anti-progestin. It binds to progesterone receptor and glucocorticoid receptor, having an affinity with progesterone receptor in rabbit endometrium five-fold higher than that of progesterone and thereby having strong anti-progesterone effect. It causes degeneration of pregnant villus tissue and decidual tissue, endogenous prostaglandin (PG) release, luteinizing hormone decrease, corpus luteum dissolution, and necrosis of embryo sac whose development depends on corpus luteum, leading to abortion. Therefore, it can be used as a non-surgical medicine for stopping early pregnancy. It can also be used, inter alia, in contraception and as an antineoplastic. (The Antiprogestin Steroid Ru486 and Human Fertility Control, 1985, New York: Plenum Press) .
-
Onapristone (11β-[4-(N,N-diemthylamino)phenyl]-17α-hydroxy-17β-(3-hydroxypropyl)-13α-4,9-estradiene-3-one), is a steroid compound which is disclosed in German Patent No. 3,321,826 to Schering AG, published Dec. 20, 1984. It has a strong antiprogestin activity and can be used in abortion (American Journal of Obstetrics and Gyencology, 1987, 157:1065-1074), anticancer (Breast Cancer Research and Treatment, 1989, 14:275-288), etc. It was reported that onapristone had toxicity to human liver (European Journal of Cancer, 1999, 35(2):214-218).
-
Lilopristone (11β-[4-(N,N-dimethylamino) phenyl]-17α-[3-hydroxy-1(Z)-propenyl]-17β-hydroxy-4,9-estradiene-3-one) is a steroid compound which is disclosed in German Patent No. 3,347,126 to Schering AG, published July 11, 1985. It has a strong antiprogestin activity and can be used in abortion, contraception (American Journal of Obstetrics and Gyencology, 1987, 157:1065-1074), etc. It was reported that the clinical effect of lilopristone in stopping early pregnancy was only equivalent to that of mifepristone (Human Reproduction, 1994, 9(1):57-63).
-
ZK112993 (11β-(4-acetylphenyl)-17α-(1-propinyl)-17β-hydroxy-4,9-estradiene-3-one) is as steroid compound which is disclosed in German Patent No. 3,504,421 to Schering AG, published Aug. 7, 1986. It has a potent antiprogestin activity and can be used in, inter alia, anticancer (Anticancer Res., 1990, 10:683-688).
-
In European Patent No. 321,010 to Akzo NV, The Netherland published June 21, 1989 are disclosed “11-arylsteroid compounds” having a strong antiprogestin activity.
PATENT
WO 2001018026
http://www.google.com/patents/EP1219632A1?cl=en

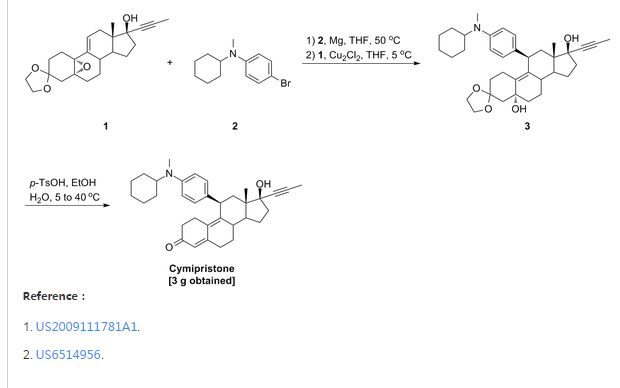
The preparation method of the present invention includes the following single- or multi-step procedures:
1. Method for the preparation of 11β-[4-(N-methyl-N-cyclohexylamino)phenyl]-17α-(1-propinyl)-17β-hydroxy-4,9-estradiene-3-one (IV) which includes the following steps:
(1) Preparation of Grignard reagent (III)

4-bromo-N-methyl-N-cyclohexylaniline (II) is reacted with magnesium in tetrahydrofuran (THF) to obtain Grignard reagent of formula (III).
(2) C11 additive reaction

Compound of formula (IV) and the Grignard reagent of formula (III) prepared in step (1) are brought to an additive reaction to obtain compound of formula (V).
(3) Hydrolytic reaction

The compound of formula (V) prepared in step (2) is subjected to a hydrolytic reaction to obtain compound of form (VI).
2. Method for preparation of 11β-[4-(N-cyclohexylamino)phenyl]-17α-(1-propinyl)-17β-hydroxy-4,9-estradiene-3-one (XI) which includes the following steps:
(1) Preparation of Grignard reagent of formula (IX)

4-bromo-N-cyclohexylaniline (VII) is first protected by trimethylchlorosilane, then reacted with magnesium in THF to obtain Grignard reagent of formula (IX).
(2) C11 additive reaction

Compound of formula (IV) and the Grignard reagent of formula (IX) prepared in step (1) are brought to an additive reaction to obtain compound of formula (X).
(3) Hydrolytic reaction

The compound of formula (X) prepared in step (2) is subjects to a hydrolytic reaction to obtain compound of formula (XI).
Example 2:
-

-
9g 4-bromo-N-cyclohexylaniline (VII) (CA registration number [113388-04-8], see Synthetic Communications, 1986, 16(13): 1641-1645 for its preparation) was placed into a four-necked flask and 15 ml (1.5 mol/L) n-BuLi solution in n-hexane. The mixture was stirred for 30 min at room temperature. Then 8 g trimethylsilyl chloride (Me3SiCl) was added and the mixture was stirred for 1 hour. Solvent and excessive Me3SiCl was evaporated under reduced pressure to yield 4-bromo-(N-cyclohexyl-N-trimethylsilylaniline) (VIII) which was formulated into a solution with 7.5 ml anhydrous tetrahydrofuran for further use.
-
1.3 g magnesium was placed into a four-necked flask and a small amount of the above solution was added dropwise and slowly at 40°C. After completion of addition, the temperature was kept for 1 hour to yield a solution of 4-(N-cyclohexyl-N-trimethylsilylamino)phenylmagnesium bromide (IX) in tetrahydrofuran for further use.
- Preparation of 11β-[4-(N-cyclohexylamino)phenyl]-17α-(1-propinyl)-17β-hydroxy-4,9-estradiene-3-one (XI)(1) Preparation of 4-(N-cyclohexyl-N-trimethylsilylamino)phenyl magnesium bromide (IX)

(2) Preparation of 3,3-ethylenedioxy-5α,17β-dihydroxy-11β-[4-(N-cylohexylamino)phenyl]-17α-(1-propinyl)-9(10)-estrene(X).

- 5g 3,3-ethylenedioxy-5,10-epoxy-17α-(1-propinyl)-17β-hydroxy-9(11)-estrene (IV) was placed into a four-necked flask and 10 ml anhydrous tetrahydrofuran and a catalytic amount of cuprous chloride (Cu2Cl2) added. Then solution of 4-(N-cyclohexyl-N-trimethylsilylamino)phenyl magnesium bromide (IX) in tetrahydrofuran was added dropwise and slowly while controlling the temperature below 5°C. After completion of addition, the mixture was allowed to react for 2 hours at room temperature and to stand overnight. Saturated ammonium chloride aqueous solution was added and the tetrahydrofuran layer separated which was washed with saturated ammonium chloride solution. The solution in tetrahydrofuran was washed with saturated saline and dried over anhydrous sodium sulfate. Evaporation of tetrahydrofuran under reduced pressure yielded a residual which was chromatographed on silica gel column using cyclohexane: acetone (5:1) as developing agent to yield 3 g 3,3-ethylenedioxy-5α,17β-dihydroxy-11β-[4-(N-cyclohexylamino)phenyl]-17α-(1-propinyl)-9(10)-estrene(X).
-
IR (KBr) cm-1: 3420 (C5, C17-OH), 1610, 1510 (benzene backbone), 840, 808 (ArH).
1H NMR (CDCl3) δ ppm: 0.52(3H, S, C13-CH3), 2.72(3H, S, N-CH3), 3.92(4H, m, -O-CH2CH2-O-), 4.24(1H, m, C11-H), 6.65-7.00 (4H, ArH).
(3) Preparation of 11β- [4- (N-cyclohexylamino)phenyl] -17α- (1-propinyl) -17β-hydroxy-4,9-estradiene-3-one (XI).

- 1.5g 3,3-ethylenedioxy-5,17β-dihydroxy-11β-[4-(N-cyclohexylamino)phenyl]-17α-(1-propinyl)-9(10)-estrene (X) and 0.75 g para-toluenesulfonic acid (PTS) were dissolved in 15 ml 90 % ethanol (v/v). The mixture was stirred for 2 hours while controlling the temperature at 40°C-50°C. After completion of the reaction, the reactant was poured into diluted sodium hydroxide aqueous solution, extracted with dichloroethane, washed with water to neutrality, and dried over anhydrous sodium sulfate. Evaporation of the solvent and chromatography on silica gel column using cyclohexane: ethyl acetate (5:1) as developing agent yielded 0.9 g 11β-[4-(N-cyclohexylamino)phenyl]-17α-(1-propinyl)-17β-hydroxy-4,9-estradiene-3-one (XI).
-
IR (KBr) cm-1: 3400 (C17-OH), 1658 (unsaturated ketone), 1613, 1514 (benzene backbone), 865, 810 (ArH).
1H NMR (CDCl3) δ ppm: 0.50 (3H, S, C13-CH3), 1.76 (3H, S, C≡C-CH3), 4.32(1H, S, C11-H), 5.75(1H, S, C4-H), 6.9-7.10 (4H, ArH).
PATENT
PATENT
Example 1
Race meters mifepristone synthetic routes:

Epoxy adduct match rice mifepristone
(N- hexylamino methylcyclohexyl) phenyl magnesium bromide (1) 4-
In the four-necked flask, 1.4 g of magnesium into pieces (Mg) and 10 ml of anhydrous tetrahydrofuran (THF), no iodine or add a little change, at about 50 ° C, a solution of 10.86 g of 4-bromo-methyl -N- cyclohexyl aniline (dissolved in 24 ml of anhydrous tetrahydrofuran) dropwise Bi, incubation was continued for 1 hour with stirring to give 4- (N- methyl-cyclohexylamino) phenyl magnesium bromide tetrahydrofuran solution (to be used in the next step an addition reaction ).
(2) 3,3-ethylenedioxy -5 α, 17 β – dihydroxy -11 β – [4- (Ν- methyl -Ν- cyclohexylamino) phenyl] -17 α – (1- propyl block-yl) -9 (10) – Preparation of estra-ene (adduct) of
In the four-necked flask, into 5 g of 3,3-ethylenedioxy-5,10-epoxy -17 α – (1- propynyl) – 17 (3 – hydroxy – 9 (11) – estra-ene (epoxy), 29.1 ml anhydrous tetrahydrofuran (THF) and 0.1 g cuprous chloride (of Cu 2 of Cl 2 ), a solution of 4- (N- methyl -N-cyclohexylamino) phenyl magnesium bromide tetrahydrofuran
Nan solution, temperature control 5. C, the drop was completed, the incubation was continued for 5 hours, the reaction was completed, the reaction solution was poured into saturated aqueous ammonium chloride solution, points to the water layer, the organic layer was washed with saturated ammonium chloride solution, the aqueous layer extracted with ethyl acetate number times, the organic layers combined, washed with saturated aqueous sodium chloride, dried over anhydrous sodium sulfate, and concentrated under reduced pressure, a silica gel column, eluent cyclohexane: acetone = (5: 1) to give 3,3-ethylene dioxo -5 α, 17 β – dihydroxy -11 β – [4- (- methyl -Ν- cyclohexylamino) phenyl] -17 α – (1- propynyl) -9 (10) – female steroidal women (adduct) solid 6 grams.
IR. ‘KBi cm- ^ SlS OI ^ ^ -OH lS jSlS benzene backbone), 819 (aromatic hydrogen). NMR Ή: (CDC1 3 ) ppm by [delta]: 0.47 (3H, the S, the C IR CH 3 ), 1.88 (3H, the S, the C ≡ the C-CH 3 ), 2.72 (3H, the S, the N-CH 3 ), 6.65- 7.03 (4H, ArH) O
(3) 11 β – [4- (N- methyl -N- cyclohexylamino) phenyl] -17 α – (1- propynyl) -17 β – hydroxy-estra-4,9-diene – Preparation of 3-one (match rice mifepristone) of
‘2.5 g of p-toluenesulfonic acid (PTS) and 5 grams of 3,3-ethylenedioxythiophene -5 α, 17 β – dihydroxy -11 β – [4- (Ν- methyl cyclohexylamino) phenyl] -17 α – (1- propynyl) -9 (10) – estra-ene (adduct) was dissolved in 50 ml of ethanol 90% (V / V), and at 5 ° C – 40 ° C the reaction was stirred 3 hours, the reaction solution was poured into dilute aqueous sodium hydroxide solution, the precipitated solid was suction filtered, washed with water until neutral, the filter cake was dissolved in 50 ml of ethyl acetate, then with saturated aqueous sodium chloride solution to the water layer was evaporated part of the solvent, the precipitated solid was suction filtered, and dried to give a pale yellow solid 11 β – [4- (Ν- -N- methyl-cyclohexylamino)] -17 α – (1- propynyl) -17 β – hydroxy estra-4,9-dien-3-one (match rice mifepristone) 3 grams.
^ Cm & lt IRCKB 1 : 3447 (the C . 17 -OH), among 1655 (unsaturated ketone), 1607,1513 (benzene backbone), 865,819 (aromatic hydrogen).
NMR ¾: (CDC1 3 ) ppm by [delta]: 0.56 (3H, the S 5 the C 13 -CH 3 ), 1.89 (3H, the S 5 -C ≡ the C-the CH3), 2.74 (3H, the S, the N-the CH3), 4.34 ( lH, the S, the C N -H), 5.75 (lH, the S, the C 4 -H), 6.68-6.99 (4H, ArH).
PATENT
PATENT
PAPER
Volume 878, Issues 7–8, 1 March 2010, Pages 719–723
Determination of cymipristone in human plasma by liquid chromatography–electrospray ionization-tandem mass spectrometry
doi:10.1016/j.jchromb.2010.01.027
Abstract
A rapid, specific and sensitive liquid chromatography–electrospray ionization-tandem mass spectrometry method was developed and validated for determination of cymipristone in human plasma. Mifepristone was used as the internal standard (IS). Plasma samples were deproteinized using methanol. The compounds were separated on a ZORBAX SB C18 column (50 mm × 2.1 mm i.d., dp 1.8 μm) with gradient elution at a flow-rate of 0.3 ml/min. The mobile phase consisted of 10 mM ammonium acetate and acetonitrile. The detection was performed on a triple-quadruple tandem mass spectrometer by selective reaction monitoring (SRM) mode via electrospray ionization. Target ions were monitored at [M+H]+m/z 498 → 416 and 430 → 372 in positive electrospray ionization (ESI) mode for cymipristone and IS, respectively. Linearity was established for the range of concentrations 0.5–100 ng/ml with a coefficient correlation (r) of 0.9996. The lower limit of quantification (LLOQ) was identifiable and reproducible at 0.5 ng/ml. The validated method was successfully applied to study the pharmacokinetics of cymipristone in healthy Chinese female subjects.
CHEMICAL ABSTRACTS, vol. 115, no. 25, 23 December 1991 (1991-12-23) Columbus, Ohio, US; abstract no. 270851g, X. ZHAO ET AL.: “Synthesis and terminating early pregnancy effect of mifepristone derivatives” page 117; XP002219009 & ZHONGGUO YAOKE DAXUE XUEBAO, vol. 22, no. 3, 1991, pages 133-136,
//////////Cymipristone, Saimisitong, NDA Filed , china, Shanghai Siniwest Pharmaceutical Chemical Technology Co., Ltd., Shanghai Zhongxi Pharmaceutical Co. Ltd., Xianju Pharmaceutical Co., Ltd,
Idarucizumab


Idarucizumab
(Praxbind®) Approved
An antidote for rapid reversal of dabigatran-induced anticoagulation indicated for emergency surgery (urgent procedures) and life-threatening or uncontrolled bleeding in patients treated with dabigatran.

BI-655075
CAS No.1362509-93-0
Other Names
- BI 655075
- Idarucizumab
- Praxbind
Protein Sequence
Sequence Length: 444, 225, 219multichain; modified (modifications unspecified)
Idarucizumab, sold under the brand name Praxbind, is a monoclonal antibody designed for the reversal of anticoagulant effects ofdabigatran.[1][2]
This drug was developed by Boehringer Ingelheim Pharmaceuticals. A large study sponsored by the manufacturer found that idarucizumab effectively reversed anticoagulation by dabigatran within minutes.[3] It was FDA approved in October 2015.[4] In the United States the wholesale cost is $3500 US.[5]
References
- Statement On A Nonproprietary Name Adopted By The USAN Council – Idarucizumab, American Medical Association.
- World Health Organization (2013). “International Nonproprietary Names for Pharmaceutical Substances (INN). Proposed INN: List 109” (PDF). WHO Drug Information 27 (2).
- Pollack, Charles V.; Reilly, Paul A.; Eikelboom, John; Glund, Stephan; Verhamme, Peter; Bernstein, Richard A.; Dubiel, Robert; Huisman, Menno V.; Hylek, Elaine M. (2015-08-06).“Idarucizumab for Dabigatran Reversal”. The New England Journal of Medicine 373 (6): 511–520. doi:10.1056/NEJMoa1502000. ISSN 1533-4406. PMID 26095746.
- “Press Announcements – FDA approves Praxbind, the first reversal agent for the anticoagulant Pradaxa”. http://www.fda.gov. Retrieved 2015-10-17.
- Elia, Joe. “Dabigatran-Reversal Agent Price Set”. Retrieved 20 October 2015.
| Monoclonal antibody | |
|---|---|
| Type | Fab fragment |
| Source | Humanized (from mouse) |
| Target | Dabigatran |
| Clinical data | |
| Trade names | Praxbind |
| Identifiers | |
| CAS Number | 1362509-93-0 |
| ATC code | V03AB37 (WHO) |
| IUPHAR/BPS | 8298 |
| ChemSpider | none |
| Chemical data | |
| Formula | C2131H3299N555O671S11 |
| Molar mass | 47.8 kg/mol |
/////Idarucizumab
Istradefylline

Istradefylline, KW-6002
(Nouriast®) Approved
A selective adenosine A2A receptor antagonist used to treat Parkinson’s disease.

KW-6002
CAS No. 155270-99-8
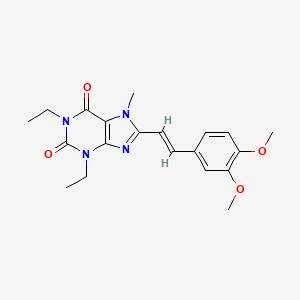
Istradefylline; 155270-99-8; KW-6002; KW 6002; 8-[(E)-2-(3,4-Dimethoxyphenyl)ethenyl]-1,3-diethyl-7-methyl-purine-2,6 -dione; (E)-8-(3,4-Dimethoxystyryl)-1,3-diethyl-7-methyl-1H-purine-2,6(3H,7H)-dione;
| Molecular Formula: | C20H24N4O4 |
|---|---|
| Molecular Weight: | 384.42896 g/mol |
Istradefylline (KW-6002) is a selective antagonist at the A2A receptor. It has been found to be useful in the treatment of Parkinson’s disease.[1] Istradefylline reduces dyskinesia resulting from long-term treatment with classical antiparkinson drugs such as levodopa. Istradefylline is an analog of caffeine.
Kyowa Hakko Kirin is developing istradefylline, a selective adenosine A2A receptor antagonist, for the once-daily oral treatment of Parkinson’s disease (PD). Adenosine A2A receptors are considered to be present particularly in the basal ganglia of the brain; the degeneration or abnormality observed in PD is believed to occur in the basal ganglia, which is recognized to play a significant role in motor control.
Commercially available dopamine replacement therapies effectively treat the early motor symptoms of PD; however, these agents are associated with development of motor complications, limiting usefulness in late stages of the disease. Istradefylline is proposed to possess a clearly distinct action site from existing agents which act on dopamine metabolism or dopamine receptors. Kyowa Hakko Kirin has received approval for istradefylline in the adjunctive treatment of PD in Japan. A New Drug Application was filed in the USA, but the FDA issued a non-approvable letter in February 2008.
PATENT
US5484920A
http://www.google.co.in/patents/US5484920
PAPER
http://www.sciencedirect.com/science/article/pii/S0960894X13003983

Scheme 1.
Synthesis of KW 6002 (2). Reagents and conditions: (i) acetic anhydride, 80 °C, 2 h, 83%; (ii) sodium nitrite, 50% acetic acid, 60 °C, 15 min, 86%; (iii) sodium dithionite, NH4OH solution (12.5% (w/v)), 60 °C, 30 min, 98%; (iv) SOCl2, toluene, 75 °C, 2 h, 97%; (v) pyridine, DCM, rt, 16 h, 66%; (vi) HMDS, cat. (NH4)2SO4, CH3CN, 160 °C, microwave, 5 h, 100% followed by (vii) MeI, K2CO3, DMF, rt, 2 h, 75%.

Synthesis
(E)-8-(3,4-Dimethoxystyryl)-1,3-diethyl-7-methyl-1H-purine-2,6(3H,7H)-dione (2)3
- J. Hockemeyer; J. C. Burbiel; C. E. Müller, J. Org. Chem. 2004, 69, 3308.
(E)-8-(3,4-Dimethoxystyryl)-1,3-diethyl-1H-purine-2,6(3H,7H)-dione (1.11 g, 3.00 mmol) was taken up in dimethylformamide (15 mL) and potassium carbonate (828 mg, 6.00 mmol). To the milky white mixture was added iodomethane (468 µL, 7.50 mmol) and it was allowed to stir at room temperature for 2 h. The mixture was then filtered and washed with water (100 mL), leaving the title compound 2 as a pale yellow solid which was dried in the oven at 110 °C (863 mg, 75%), mp: 192 °C (lit.3 191 °C). 1H NMR (400 MHz, CDCl3) δ 7.73 (d, J = 15.7 Hz, 1H), 7.18 (dd, J = 8.4, 1.9 Hz, 1H), 7.09 (d, J = 1.9 Hz, 1H), 6.90 (d, J = 8.4 Hz, 1H), 6.76 (d, J = 15.7 Hz, 1H), 4.21 (q, J = 7.1 Hz, 2H), 4.12 – 4.04 (m, 5H), 3.95 (s, 3H), 3.93 (s, 3H), 1.39 (t, J = 7.1 Hz, 3H), 1.26 (t, J = 7.0 Hz, 3H). 13C NMR (101 MHz, CDCl3) δ 155.0 (C), 150.8 (C), 150.4 (C), 150.3 (C), 149.2 (C), 148.2 (C), 138.1 (CH), 128.6 (C), 121.2 (CH), 111.2 (CH), 109.5 (CH), 109.3 (CH), 108.0 (C), 55.98 (CH3), 55.97 (CH3), 38.4 (CH2), 36.3 (CH2), 31.5 (CH3), 13.43 (CH3), 13.39 (CH3). LCMS: m/z (ESI 20 V) 385.2 (MH+, 100).
PATENT
http://www.google.com/patents/CN103254194A?cl=en
Specific synthetic route is as follows:

the above reaction is a synthetic Parkinson’s disease clinical drug KW-6002 against a yield of 83%.
Example 26 (a new synthetic method for anti-Parkinson’s disease in clinical drug KW-6002):
In addition to use in place of 3,4-dimethoxy-styryl boronic acid (0.4mmol, i.e., in formula IV, R5 is 3,4_-dimethoxy-styryl) benzene boronic acid in Example 23 and 1,3 – two-ethyl-8-phenylthio-9-methyl-xanthine (0.4mmol, i.e., Formula I, R1 is methyl, R2 and R3 are ethyl, R4 is a phenyl group) in place of Example 23 in 1 , 3,9-trimethyl xanthine -8- phenylthio, the remaining steps in Example 23 to give a white solid, yield 83%, mp = 101~103 ° C I1H NMR (⑶CI3, 600MHz): δ 7.71 (d, J = 15.6Hz, 1H), 7.17 (dd, J = 8.2,1.9Hz, 1H), 7.07 (d, J = L 9Hz, 1H), 6
• 88 (d, J = 8.2Hz, 1H), 6.74 (d, J = 15.8Hz, 1H), 4.19 (q, J = 7Hz, 2H), 4.07 (q, J = 7Hz, 2H), 4.03 (s , 3H), 3.93 (s, 3H), 3.90 (s, 3H), 1.36 (t, J = 7Hz, 3H), 1.23 (t, J = 7Hz, 3H); 13C NMR (150MHz, CDCl3): 155.1, 150.8,150.4,150.2,149.2,148.2,138.2,128.6,121.2, 111.2,109.5,109.3,108.0,56.0,55.9,38.4,36.3,31.5,13.4,13.4; HRMS: calcd for C20H25N4O4 (M + H) +385.187
6, Found385.1879. It indicates that the white solid was 8- (3,4-dimethoxy-styryl) structural formula shown KW-6002 (E) -1,3_ diethyl-7-methylxanthine.

In contrast, KW-6002 is a new drug to treat Parkinson’s disease developed by Kyowa Hakko in Japan, Japan and the United States is currently the second phase of clinical trials. Literature (. J.Hockemeyer, JCBurbiel andC.E.Muller, J.0rg.Chem, 2004,69,3308) through the following synthetic route:

The synthetic route requires five steps, with a total yield of 33%, and there is the use of environmentally unfriendly halogenated solvent methylene chloride, the reaction requires high pressure high temperature (170~180 ° C) and other shortcomings. By comparison, the present invention starting from 8- phenylthio xanthine coupling reaction catalyzed by palladium simple, a yield of 83% was synthesized KW6002, it is currently the most efficient synthesis route KW-6002’s. In particular, the multi-step synthesis route to avoid the complex operation of the reactor, but under relatively mild conditions (60 ° C) conduct, simple operation, suitable for scale synthesis.
PATENT
http://www.google.com/patents/CN104744464A?cl=en
itraconazole theophylline (Istradefylline, KW6002), the chemical name 8 – [(E) -2- (3, 4- dimethoxyphenyl) ethenyl] -1,3-diethyl -7 – methyl-purine-2,6-dione, CAS number: 155270-99-8, structural formula shown below.

itraconazole Theophylline is a selective adenosine A2a receptor antagonist, by changing the activity of neurons in Parkinson’s disease patients to improve motor function, for the treatment of Parkinson’s disease and Parkinson’s disease improve early dyskinesia.
The invention and JPH0940652A European Patent 0,590,919 discloses a method for preparing itraconazole and theophylline. WO 2004/099207 published good solubility stability of a particle size of less than 50 micrometers 8 – [(E) -2- (3, 4- dimethoxyphenyl) ethenyl] -1,3- diethyl-7-methyl-purine-2,6-dione crystallites.
Example 1 Preparation of theophylline itraconazole Example

ships equipped with a mechanical stirrer, a thermometer, a 2L 4-neck flask was added 30g8 – [(E) -2- (3, 4- dimethoxyphenyl) ethenyl] -1,3-diethyl- -7- hydrogen – purine-2,6-dione (Intermediate A), 400mL N, N- dimethylformamide and 15g of potassium carbonate, and 25g of methyl iodide and heated to 80 ° C after the reaction was stirred 8h, added 200mL water, cooled to room temperature, and stirring was continued crystallization 2h. The resulting suspension was suction filtered, washed with water after the cake was 800mL sash, 50 ° C under blast drying 24h, 32g give a pale yellow solid, for each polymorph of itraconazole theophylline preparation example the following examples.
References
- Peter A. LeWitt, MD, M. Guttman, James W. Tetrud, MD, Paul J. Tuite, MD, Akihisa Mori, PhD, Philip Chaikin, PharmD, MD, Neil M. Sussman, MD (2008). “Adenosine A2A receptor antagonist istradefylline (KW-6002) reduces off time in Parkinson’s disease: A double-blind, randomized, multicenter clinical trial (6002-US-005)”. Annals of Neurology 63 (3): 295–302. doi:10.1002/ana.21315. PMID 18306243.
Reference:1. EP0590919A1.
2. US5484920A.
3. US5543415A.
4. J. Org. Chem. 2004, 69, 3308-3318.
5. Bioorg. Med. Chem. Lett. 1997, 7, 2349-2352.
6. Bioorgan. Med. Chem. 2003, 11, 1299-1310.
7. Bioorg. Med. Chem. Lett. 2013, 23, 3427-3433.
8. Chinese Journal of Pharmaceuticals 2010, 41, 241-243.
9. JP0940652A.
10. Org. Biomo. Chem. 2010, 8, 4155-4157.
1. Chem. Commun. 2012, 48, 2864-2866.
2. CN103254194A.
| CN104744464A * | Nov 15, 2013 | Jul 1, 2015 | 南京华威医药科技开发有限公司 | Istradefylline crystal forms |
-
Istradefylline Systematic (IUPAC) name 8-[(E)-2-(3,4-dimethoxyphenyl)vinyl]-1,3-diethyl-7-methyl-3,7-dihydro-1H-purine-2,6-dioneIdentifiers CAS Number 155270-99-8 
ATC code none PubChem CID 5311037 IUPHAR/BPS 5608 ChemSpider 4470574 UNII 2GZ0LIK7T4 KEGG D04641 ChEMBL CHEMBL431770 Chemical data Formula C20H24N4O4 Molar mass 384.429 g/mol

//////Istradefylline, KW-6002, Nouriast®, Approved, A selective adenosine A2A receptor antagonist, Parkinson’s disease,
O=C2N(c1nc(n(c1C(=O)N2CC)C)\C=C\c3ccc(OC)c(OC)c3)CC
GMP Oversight of Medicines Manufacturers in the European Union


A System of Equivalent Member States, a Coordinating Agency and a Centralized Institution
The regulatory system for supervision of pharmaceutical manufacturers and GMP inspection in the European Union is one of the most advanced in the world. Due to the globalization of pharmaceutical manufacture, it also affects industry, regulators and patients outside the European Union. This system, however, is often poorly understood beyond the EU borders.
What follows is an explanation of the EU system in order to increase awareness and facilitate cooperation on GMP between European Union regulators and those outside the European Union.

The European Union
The European Union includes 28 Member States located in Europe, which are: Austria, Belgium, Bulgaria, Croatia, Cyprus, Czech Republic, Denmark, Estonia, Finland, France, Germany, Greece, Hungary, Ireland, Italy, Latvia, Lithuania, Luxemburg, Malta, Netherlands, Poland, Portugal, Romania, Slovakia, Slovenia, Spain, Sweden, and United Kingdom. The EU total population is about 500 million people.
View original post 4,770 more words
GMP/GDP: When will I be inspected by the Authorities?
Various competent authorities are performing inspections. But who is subject to such an inspection?
GMP Inspections are carried out at Manufacturer Licence Holders
A manufacturer of medicinal products must meet Good Manufacturing Practice (GMP) standards. These standards are defined in various laws and regulations. In the EU the compliance with these regulations is checked and assessed by the national competent authorities. The overall goal is to have medicinal products of consistent high quality that meet the requirements of the marketing authorisation (MA) or product specification.
If a company supplies product to the USA, the U.S. Food and Drug Administration (FDA) might inspect the site assuring that drugs, medical devices, certain active pharmaceutical ingredients (APIs) and biological products manufactured in foreign countries and intended for U.S. distribution are in compliance with the applicable U.S. law and regulations.
GDP Inspections are carried out at Wholesale Dealer Licence Holders
Good Distribution Practice…
View original post 312 more words
1R,2S-Methoxamine

1R,2S-methoxamine, also known as L-erythro-methoxamine
CAS 13699-29-1
- Molecular Weight, 211.26, C11 H17 N O3
HYDROCHLORIDE
(1R,2S)-isomer HCl salt of 1 -(2,5-dimethoxyphenyl)-2-amino-1 -propanol also called as (1R, 2S)methoxamine hydrochloride
CAS 16122-04-6
Used as a pressor agent, as a vasoconstrictor, as a nasal decongestant, in ophthalmology and also found very effective in the treatment of faecal incontinence.
treatment of relief of fecal incontinence and anal itch (pruritis ani) , particularly for patients who have had a major bowel resection and reanastomosis .
Anal or fecal incontinence is the inability to voluntarily control the passage of feces or gas through the anus. It may occur either as fecal soiling or as rare episodes of incontinence for gas or watery stools. It is a very distressing condition that can result in self-inflicted social isolation and despair.
Conventional treatments for fecal incontinence include drug therapy to improve stool consistency, such as morphine, loperamide and codeine phosphate to reduce gut motility, and laxatives to soften stools and relieve constipation. Biofeedback training is another treatment which involves muscle strengthening exercises to improve anal canal resting pressure, and squeeze pressure, and to teach symmetry of anal canal function. The most common form of treatment however, is surgical repair, such as the creation of a neo-sphincter which involves grafting on muscle from other parts of the anus, or a colostomy. (Gastroenterology in Practice, Summer 1995, pl8- 21; Dig Dis 1990; 8:179-188; and The New England Journal of Medicine, April 1992, pl002-1004) . In mild cases of anal leakage, the patient will often try and plug the anus with a ball of cotton wall.
In Gut, 1991, 32, p.345-346 it was reported that two thirds of patients with idiopathic faecal incontinence had a decreased anal resting pressure resulting from an abnormal internal sphincter function. In many incontinent patients, the internal anal sphincter was found to be abnormally thin, while others had an external anal sphincter defect. It has also been reported that in vi tro contractile response of the internal anal sphincter to noradrenaline is decreased in incontinence, (Br. J. Surg. 1992, vol 79, August, p829-832; Digestive Diseases and Sciences, vol 38, no. 11, Nov. 1993, pl961-1969) . A further discussion of the innervation and control of the internal anal sphincter and drugs which can increase or decrease the normal anal resting pressure, is discussed in the text book Coloproctology and the Pelvic Floor (Butterworths) , second edition, 1992, at chapter 3 p37-53; Automic Control of Internal Anal Sphincter; and Journal of Clinical Investigation 1990, 86: p424-429.
In Surgery 1990; 107: p311-315 sodium valproate was found to be useful in the treatment of minor incontinence after ileoanal anastomosis.
It has now surprisingly been found that fecal incontinence and anal itch can be resolved by treatment with α adrenergic agonists, nitric oxide synthase inhibitors, prostaglandins F2α, dopamine, morphine, β-blockers such as propranolol, and 5-Hydroxytryptamine (5-HT) .
This is surprising since it was always thought that once an anal sphincter began functioning abnormally, the patient would require major surgery.
In this way the anal leakage is reduced or eliminated without the patient having to undergo major surgery.
Accordingly in a first aspect of the invention there is provided use of a physiologically active agent selected from an α adrenergic agonist, nitric oxide synthase inhibitor, prostaglandin F2α, dopamine, morphine, β-blockers, and 5- Hydroxytryptamine in the preparation of a medicament for the treatment or prophylaxis of fecal incontinence or anal itch.
The agents of the invention appear to at least partially treat the incontinence by increasing the resting pressure of the internal anal sphincter. Preferred agents are λ adrenergic agonists, nitric oxide synthase inhibitors, and prostaglandins F2α.
Examples of suitable aλ adrenergic agonists are nor- adrenalin, methoxamine, but particularly preferred is phenylephrine .
Examples of suitable F2α prostaglandin are dinoprost and carboprost.
Examples of suitable NO synthase inhibitors are
NG-monnoommeetthhyyll–LL–aarrggiinn:ine (L-NMMA) , and NG-nitro-L-arginine methyl ester ( -NAME)
The medicament can contain a single active agent or a combination of any of the above active agents.
Nitric Oxide (NO) synthase inhibitors such as LNMMA have previously been suggested for the therapeutic treatment of septic shock.
The prostaglandins, along with thromboxanes and leukotrienes are all derived from 20 -carbon polyunsaturated fatty acids and are collectively termed eicosanoids. F2α prostaglandins are derived in vivo from the endoperoxide prostaglandin H2which is in turn derived from leukotrienes. Clinically, F2α prostaglandins such as dinoprost and carboprost are used as uterine stimulants in the termination of pregnancy, missed abortion or the induction of labour.
Phenylephrine (an αx adrenergic agonist) is used as a mydriatic in ophthalmology, and as a decongestant , for example, in cold and flu remedies.
However there has been no suggestion to the inventors knowledge of using any of these active agents to treat fecal incontinence or anal itch. As used herein “fecal incontinence” includes all types of anal leakage from minor leakage or ‘spotting’ through moderate leakage, to major instances of faecal incontinence, and includes neurogenic, active, urge and passive incontinence.
More particularly the class of incontinent patients who will benefit most from the present invention are those with idiopathic incontinence and those whose incontinence is at least partly due to a weakness of either the internal or external anal sphincter, especially those with a normal or low maximum anal pressure and a structurally intact internal anal sphincter muscle, such as with an abnormally thin sphincter. However patients with minor structural damage such as a fragmented sphincter would still benefit from the invention. Not only incontinent patients with a damaged or abnormal internal sphincter can be treated, but also patients with a damaged or abnormal external sphincter since the increase in the internal anal resting tone induced by the invention will compensate for a poorly functioning external sphincter.
Another class of patients who particularly benefit from the invention are post-surgical patients who have had major bowel resection and reanastomosis . For example patients with ileoanal pouch (restorative proctocolectomy) , coloanal (with or without colonic pouch) anostomosis, lower anterior resection, and colectomy with ileorectal anastomosis.
The damage to the sphincter could be caused by trauma, such as experienced in child birth, surgical operations, or road traffic accidents. Furthermore it is also believed that incontinence caused by primary internal anal degeneration can also be relieved by the invention.
Anal leakage also often leads to pruritis of the anus and therefore by reducing or eliminating the leakage, the pruritis or anal itch is also relieved or prevented. Furthermore, as a result of the increased anal resting pressure, the patient no longer has the discomfort of distended anal sphincter muscles.
Methoxamine contains two chiral carbons and thus exists in four isomeric forms. Of all the isomeric forms, the studies revealed (1R,2S)- isomer to be therapeutically active.
US patent 2359707 describes the process for the synthesis of racemic β-(2,5-dimethoxy phenyl)-P-hydroxy-isopropyl amine in neutral, acid salt and its derivative from 2,5- dimethoxy propiophenone by treatment with methylnitrite in diethyl ether medium to obtain 2,5-dimethoxy-a-isonitrosopropiophenone hydrochloride. It is further reduced with palladium on carbon to yield β-(2,5-dimethoxyphenyl)-p-ketoisopropylamine hydrochloride and then with platinum black to get p-(2,5-dimethoxyphenyl)-β- hydroxyisopropyl amine hydrochloride. The described process for di-methoxamine HC1 is not cost-effective, due to the use of two expensive catalysts (platinum black and palladium carbon), solvent diethyl ether and involves more number of steps. The other drawback being it is racemic mixture and cannot be used directly as drug. The process described did not specify the quality of the product.
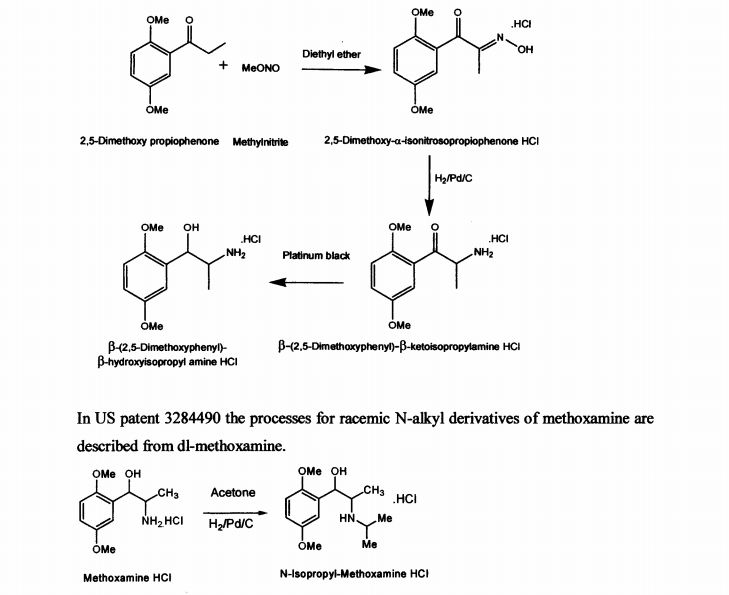
In US patent 3284490 the processes for racemic N-alkyl derivatives of methoxamine are described from dl-methoxamine.
JP 63165348 describes process for production of optically active l-(2,5- dimethoxyphenyl)-2-aminophenol by resolving racemic compound with the use of optically active L-N-acetylleucine as resolving agent. The disadvantages of the process are less yield, low quality and use of expensive naturally occurring amino acid, which prevents from employing this method on commercial scale.
WO 03/055474 A1 discloses mainly, the use of (1R, 2S)-methoxamine in the treatment of faecal incontinence at low doses without local or systemic side effects when used topically. The patent also described the synthesis of (1R, 2S)-methoxamine, from L- alanine, by protecting the amino group using methylchloroformate, converting carboxy
group of the N-protected alanine into an acid chloride insitu followed by reaction with an amine to produce an N-protected (S)-alanine amide and coupling that compound with a brominated 2,5-dimethoxybenzene in the presence of n-butyllithium or a magnesium based reagent to give (S)-amino-l-(2,5-dimethoxy-phenyl)-l-propanone, the amino group of which is protected .The reduction of the N-protected propanone was carried out using dimethylphenylsilane and the protecting group was removed by treatment with potassium hydroxide. Other method adopted in the patent to isolate (1R,2S)methoxamine is by separation of racemic methoxamine using chiral column.
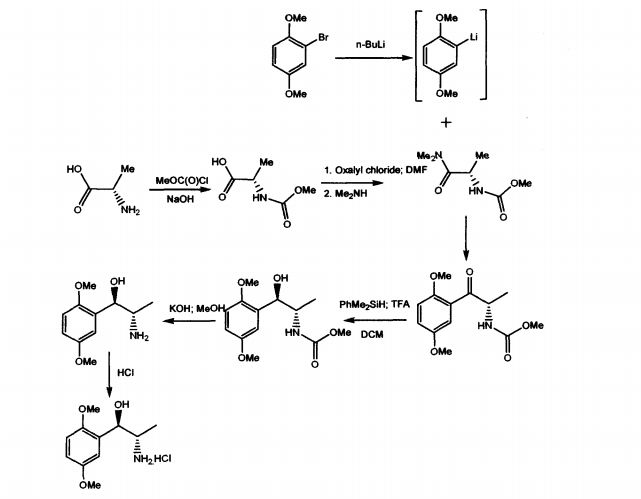
The prior art suffers with some of the disadvantages like using n-butyllithium, which is pyrophoric, expensive and causes hazards to commercial scale. Also, the separation of racemic Methoxamine using chiral column mentioned in the patent can be considered for
isolating small quantities of the required isomer for analytical purposes but cannot be adopted on commercial scale for production of the drug.
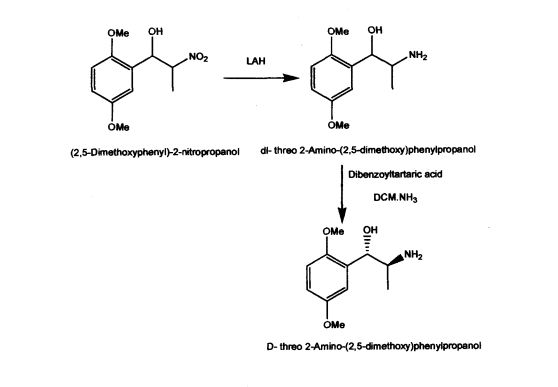
US Patent 5962737 described stereospecific synthesis of the racemic threo isomers of 2- nitro-1 -phenylpropanols by reacting benzaldehyde derivative with nitroalkane in the presence of a tertiary amine and reducing 2-nitro-l-phenylpropanols with lithium aluminium hydride to 2-amino-l-phenylpropanols. Also described is phase transfer resolution of racemic mixtures of 2-amino-l-phenylpropanol and its derivatives into their optically pure isomers by reacting with the mono alkali metal salt of tartaric acid ester in a two phase system of a hydrocarbon and water. The specification further describes optically pure isomer D-threo 2-amino-( 1 -dialkoxy or alkoxy)phenylpropanol by resolution of dl- threo 2-amino-( 1 -dialkoxy or alkoxy)phenylpropanol by using dibenzoyltartaric acid. The synthesis of the product (lS,2S)-threo 2-amino-(l-dialkoxy or alkoxy) phenyl propanol involves the use of expensive and hazardous chemicals like LAH making the process technically and commercially difficult for implementation.
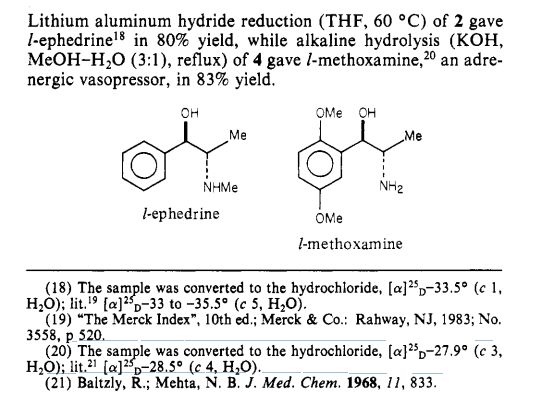
Paper
Journal of the American Chemical Society (1984), 106(16), 4629-30
http://pubs.acs.org/doi/pdf/10.1021/ja00328a062
PATENT
http://www.google.com/patents/EP2275099A1?cl=en
EXAMPLE 3Synthesis of 1R,2S-Methoxamine(S)-N-Methoxycarbonyl alanine
To a stirred solution of L-alanine (300g, 3.37 mol sodium hydroxide (1N, 1800 cm3) at 0°C in an ice bath was added dropwise, over 2 hours, methyl chloroformate (274 cm3, 3.54 mol). The pH of the solution was maintained at 9 by the addition of sodium hydroxide (5N). The reaction mixture was stirred at 0°C for 3 hours whereupon it was acidified to pH 1 by the addition of phosphoric acid solution (15%) and extracted with diethyl ether (5 x 1000 cm3). The combined organic extracts were dried (MgSO4) and concentrated under reduced pressure to yield the product as a viscous green oil (386 g, 78%). 1H NMR (250 MHz; C2HCl3) 1.48 (3H, d, J7.25, CH3), 3.72 (3 H, s, COCH3), 4.40 (1 H, quintet, J7.25, CH), 5.31 (1 H, bs, NH).
(S)-N-Methoxycarbonyl alaninedimethylamide
To a stirred solution of MeOC-alanine (227 g, 1.54 mol) and dimethylformamide (DMF) (25 cm3) in dry dichlorourethane (DCM) (2000 cm3) at 0°C was added dropwise oxalyl chloride (146 cm3, 1.62 mol) over a period of 2 hours. The solution was stirred at 0°C until the evolution of gasses ceased whereupon a basic solution of dimethylamine (676 g, 7.70 mol) in NaOH (3 N, 2000 cm3) was added. The aqueous layer was extracted with diethyl ether (2 x 500 cm3) and the combined organic layers dried (MgSO4) and concentrated under reduced pressure to give the product as a white crystalline solid which required no further purification (230 g, 86%). 1H NMR (250 MHz; C2HCl3) 1.33 (3 H, d, J6.75, CH3), 2.99 3 H, s, OCH3) 3.08, (3 H, s, OCH3), 3.66 (3 H, s, COCH3), 4.66 (H, quintet, J7.00, CH), 5.75 (1 H, d, J5.75, NH).
(S)-2-[(Methoxycarbonyl)amino]-1-(2,5-dimethoxyphenyl)-1-propanone.
To a THF (1000 cm3) solution of bromo-2,5-dimethoxybenzene (55 g, 0.25 mol) at -20°C under nitrogen was addedn-butyl lithium (100 cm3, 2.5 M in hexanes, 0.25 mol). The mixture was stirred at -20°C for 0.75 hours, whereupon a THF (100 cm3) solution of amide (30 g, 0.17 mol) was added via cannula. The solution was stirred at -20°C for 2 hours and was then allowed to warm to room temperature over 1 hour and quenched by the addition of ammonium chloride solution (700 cm3). The solution was diluted with diethyl ether (1000 cm3) and the organic layer was dried (MgSO4) and concentrated under reduced pressure to give a yellow oil. The product was purified by dry flash chromatography on silica (eluant 4:1 hexane/ethyl acetate then 3:2 hexane/ethyl acetate) to give the product as a white crystalline solid (45 g, 98%). 1H NMR (250 MHz; C2HCl3) 1.36 (3 H, d, J7.0, CH3), 3.70 (3 H, s, COCH3), 3.82 (3 H, s, OCH3), 3.92 (3 H, s, OCH3), 5.43 (1 H, quintet, J 7.3, H-2), 5.80 (1 H, bs, NH), 6.94 (1 H, d, J 9.0, ArH), 7.10 (1 H, dd, J 9.0, 3.3, ArH), 7.32 (1 H, d, J 3.3, ArH).
(1R,2S)-2-[(Methoxycarbonyl)amino]-1-(2,5-dimethoxyphenyl)-1-propanol.
To a stirred solution of ketone i.e. (S)-2-[(methoxycarbonyl)amino]-1-(2,5-dimethoxyphenyl)-1-propanone (20 g, 74.9 mmol) and dimethylphenyl silane (10.7 g, 78.6 mmol) in dry DCM (500 cm3) at 0°C in an ice bath was added dropwise trithioroacetic acid (TFA) (50 cm3). The solution was stirred at 0°C for 1 h and then quenched by the addition of sodium hydroxide (500 cm3, 1 N). The organic layer was dried and concentrated under reduced pressure to give a yellow oil which solidified on standing. This solid was crystallized from ether/hexane to give the product as a white crystalline solid (15.6 g, 75%).1H NMR (250 MHz; C2HCl3) 1.03 (3 H, d, J7.0, CH3), 3.04 (1 H, d, J4.3, OH), 3.68 (3 H, s, COCH3), 3.78 (3 H, s, OCH3), 3.80 (3 H, s, OCH3), 3.94-3.99 (1 H, m, H-2), 5.05-5.15 (2 H, m, H-1 and NH), 6.72-6.85 (2 H, m, ArH) 6.97 (1 H, d, J 2.0, ArH).
(1,R,2S)-Methoxamine.
To a stirred solution of methoxycarbonyl (MeOC) protected alcohol i.e. (1R,2S)-2-[(methoxycarbonyl)amino]-1-(2,5-dimethoxyphenyl)-1-propanol (4.0 g, 14.9 mmol) in methanol (175 cm3) was added a solution of KOH (4.06 g, 72.8 mmol in water (60 cm3). The solution was cooled and acidified with phosphoric acid (15% v/v). The solution was extracted with DCM (2 x 50 cm3) and the aqueous layer basified by the addition of K2CO3. The aqueous layer was extracted with diethyl ether (5 x 50 cm3) and the combined ethereal extracts dried (MgSO4) and concentrated under reduced pressure to give the product as a clear yellow oil (1.9 g, 61%), 1H NMR (250 MHz; C2HCl3) 0.84 (3 H, d, J 7.0, CH3), 3.19-3.22 (1 H, m, H-2), 3.71 (6 H, s, 2 x OCH3), 4.67 (1 H, d, J 5.0, H-1), 6.66-6.72 (2 H, m, ArH), 6.92 (1 H, d, J 2.5, ArH).
(1R, 2S)-Methoxamine hydrochloride.
To an ice cooled solution of (1R,2S)-methoxamine (1.9 g, 9.00 mmol) in anhydrous diethyl ether (30 cm3) was passed a stream of dry HCl gas for 45 mins. The resultant precipitate was filtered by suction, washed with cold diethyl ether and dried under nitrogen to yield the title compound as a white solid. (1.5 g, 68%). 1H NMR (250 MHz; [C2H3]2SO) 0.89 (3 H, d, J 6.8, CH3), 3.37-3.42 (1 H,m,H-2), 3.71 (3 H, s, OCH3), 3.75 (3 H, s, OCH3), 5.12 (1 H, s, H-1), 5.92 (1 H, d, J 4.3, OH), 6.84 (1 H, dd, J 8.8, 3.0, ArH), 6.92-7.00 (2 H, m, ArH); HPLC.
Analytical Method for the Analysis of Methoxamine
The following method was used to analyse methoxamine samples.
Method
-
Column : Cyclobond I RSP 250 x 4.6 mm Column temperature : 23°C Mobile phase : 0.1% Tetraethylammonium pH 4.1* 95%v/v : Acetonitrile 5%v/v Flow rate : 0.6 ml/min Solution
Concentration :5 mg/l Injection volume : 2.5 µl to 20 µl Detection : UV 230 nm *Tetraethylammonium acetate pH 4.1 was prepared fresh daily.
Example 2 above allows the complete assignment of the methoxamine isomers as shown below:


PATENT
INDIAN 1020/CHE/2011
BY


The Managing Director of Malladi Drugs & Pharmaceuticals, Prashant Malladi (left), with the Chief Executive Officer, V. N. Gopalakrishnan

V.N Gopalakrishnan
CEO at Malladi Drugs & Pharmaceuticals Ltd

Prabhakaran Ranganathan
Vice President (Operations) at Malladi Drugs and Pharmaceuticals Limited

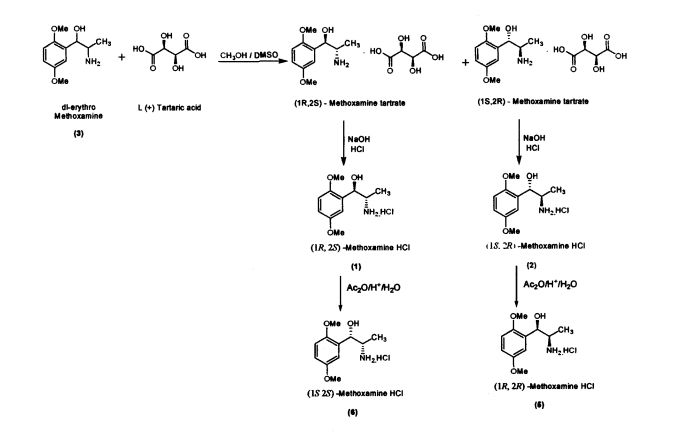
The present invention further provides an improved process for the preparation of (JS, 2S)-Methoxamine HC1 of formula (6) from (1R, 2S)-methoxamine by treating with acetic anhydride in toluene medium followed by acid hydrolysis and basification to obtain (IS, 2S)-Methoxamine base which is further acidified to form (1S,2S)- Methoxamine HC1 (6).
The present invention further provides an improved process for the preparation of (1R, 2R)-Methoxamine HC1 of formula (5) from its diastereomer (1S, 2R)-methoxamine HC1 of formula (2) by treating with acetic anhydride in toluene medium followed by acid hydrolysis and basification to obtain (1R, 2R)-Methoxamine base which is further acidified to form (1R, 2R)-Methoxamine HC1 (5).
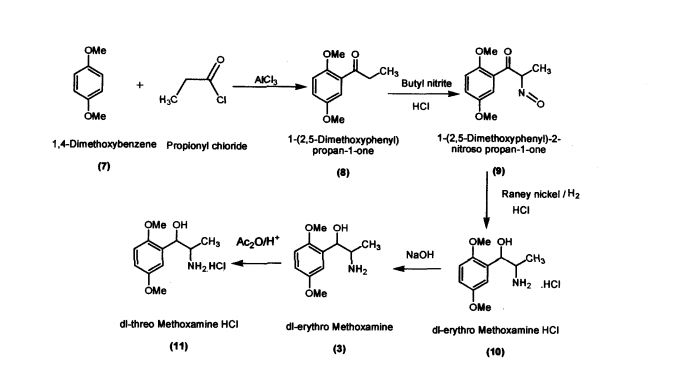
The following examples illustrate the invention.
EXAMPLES
Example 1
Preparation of l-(2,5-Dimethoxyphenyl)propan-l-one (8)
Aluminium chloride (127.4 g; 0.955 mol) was added to dichloromethane (420 mL) in a round bottomed flask under nitrogen atmosphere. The reaction mixture was cooled to -5 °C; 1,4-dimethoxybenzene (100 g; 0.724 mol) was added slowly within 15-30 minutes. Propionic chloride (87 g; 0.94 mol) dissolved in dichloromethane (245 mL) was added slowly within 2 hours. The reaction mass was allowed to stir for 2 hours and then was quenched in crushed ice (1 kilo) and HC1 (75 mL) at 0 – 5 °C. Separated the layers and the organic layer was washed with 5% sodium hydroxide solution, dried and concentrated (140 g; colorless liquid); Purity by HPLC : 99.04%
Spectroscopic interpretation
The structure of the product, l-(2,5-Dimethoxyphenyl)propan-l-one was confirmed with the help of the following spectroscopic data.
a) IR (cm-1) (KBr)
Aromatic C-H stretch at 3071, aliphatic C – H stretch at 2938, C = O stretch at 1674, benzenoid bands at 1609 and 1584, C – O stretch at 1223, C – H out of plane bending of tri-substituted benzene ring at 814,719.
b) 1H NMR(CDCb, 300 MHz) (δH)
1.16 (3H, t, -CH2-CH3), 3.0 (2H, q, -CH2-CH3), 3.78 (3H, s, -OCH3), 3.85 (3H, s, -OCH3), 6.83 – 7.72 (3H, m, aromatic protons)
c) 13C NMR (CDCb, 300 MHz) (δC)
8.44 (-CH2-CH3), 37.03 (-CH2-CH3), 55.74 (-OCH3), 56.01 (-OCH3), 113.09 – 153.41 (aromatic carbons), 202.96 (C=O)
d) Mass spectrum (ESI, methanol)
[M+Na]+ at m/z 217 (9), [M+H]+ at m/z 195 (100).
Example 2
Preparation of l-(2,5-Dimethoxyphenyl)-2-nitrosopropan-l-one (9) l-(2,5-Dimethoxyphenyl)propan-l-one (100 g; 0.515 mol) was added to dichloromethane (660 mL) in a round bottomed flask under nitrogen atmosphere. Butylnitrite (46.6 g; 0.52 mol) was slowly added in about 30 minutes at 30 – 35 °C. Diethyl ether (60.2 mL) was added to the reaction mixture and dry HC1 gas was purged for about 4 hours at 30 – 35 °C. The reaction mass was maintained for 12 hours and then concentrated under vacuum The residue obtained (60 g; Pale yellow crystalline powder); Purity by HPLC: 99.81%; mp: 104-107 °C
Spectroscopic interpretation
The structure of the product, l-(2,5-Dimethoxyphenyl)-2-nitrosopropan-l-one was confirmed with the help of the following spectroscopic data
a) IR (cm1) (KBr)
O-H stretch at 3250 (broad), aromatic C-H stretch at 3024, aliphatic C – H stretch at 2934, C = O stretch at 1688, C = N stretch at 1645, benzenoid bands at 1589 and 1504, C-O stretch at 1231, C-H out of plane bending of tri-substituted benzene ring at 745,702.
b) 1H NMR(CDCb, 300 MHz) (δh)
2.07 (3H, s, -C-CH3), 3.72 (3H, s, -OCH3), 3.76 (3H, s, -OCH3), 6.84-6.99 (3H, m, aromatic protons), 8.89 (1H, bs, OH)
c) 13C NMR (CDCb, 300 MHz) (δC)
9.16 (-C-CH3), 55.81 (-OCH3), 56.34 (-OCH3), 113.09 – 153.27 (aromatic carbons), 157.07 (C=N-OH); 193.32 (CO)
d) Mass spectrum (ESI, methanol) [M+H]+ at m/z 224 (100)
Example 3
Preparation of dl-erythro-methoxamine HC1 (10)
Raney nickel (50 g); iso-propyl alcohol (250 mL) were added to the autoclave. l-(2,5- Dimethoxyphenyl)-2-nitrosopropan-1 -one (100 g; 0.448 mol) was added slowly at 50 – 55 °C by simultaneously purging the flask with hydrogen at 2-3 Kilo pressure. When hydrogen consumption ceases, the catalyst was filtered and the filtrate was concentrated. iso-Propyl alcohol (200 mL) was added to the concentrated mass followed by acidification with HC1 to obtaindl-erythro-methoxamine HC1 (70 g; white crystalline solid)
Spectroscopic interpretation
The structure of the product, dl-erythro-methoxaxmne HC1 was confirmed with the help of the following spectroscopic data.
a) IR (cm1) (KBr)
O-H stretch at 3409, aromatic C-H stretch at 3010, aliphatic C – H stretch at 2914, HN-H str. at 2574 and 2467, benzenoid bands at 1615 and 1569, C-N stretch at 1279, C-O stretch at 1216, C-H out of plane bending of 1,2,4-tri- substituted benzene ring at 812.
b) 1H NMR (DMSO-d6, 300 MHz) (δH)
1.0 (3H,d, -CH-CH3), 3.74 (3H, s, -OCH3), 3.77 (3H, s, -OCH3), 4.89 (1H, q, -CH-CH3),6.1 (1H, d, -CH-OH), 6.87-7.01 (3H, m, aromatic protons), 8.06 (3H, bs, HN-H) The -OH proton appears to have exchanged with the solvent.
c) 13C NMR (DMSO-d6, 300 MHz) (δc)
14.75 (-CH-CH3), 52.12 (-OCH3), 55.70 (-OCH3), 55.70 (-CH-CH3), 67.25 (CH-OH), 111.89 – 153.16 (aromatic carbons)
d) Mass spectrum (ESI, methanol)
[M+H)+ at m/z 212 (100), [M-H2O]+ at m/z 194 (56).
Example 4
Preparation of(JR,2S)-Metboxamine HC1 (1) and (1S, 2R)-Methoxamine HC1 (2) dl-erythro-methoxamine HC1 (117g; 0.47 mol) was dissolved in water (350 mL) at 30-35 °C. The clear solution obtained was basified using 50% sodium hydroxide solution. dl-erythro-Methoxaumne (3) was extracted into dichloromethane (150 mL) and concentrated. Mixture of methanol/DMSO (4:1; 1650 mL) was added and the mass was heated to 50 °C. L-(+)-Tartaric acid (71.1g; 0.47mol) was added slowly and the temperature of the mass was further raised to 70 °C for complete dissolution. The mass was cooled to 35 °C and maintained for 48 hours. (IR,2.S)-Methoxamine tartrate complex (80 g) precipitated was filtered. From the filtrate on concentration was obtained (1S,2R)- methoxamine tartrate complex (82 g) (IR,25)-Methoxamine tartrate complex was added to water (250 mL) at 35 °C, basified to 12 – 13 pH with 50% sodium hydroxide solution. Dichloromethane (200 mL) was added and stirred for 30 min. Separated the org layer, dried over sodium sulphate and concentrated completely under vacuum at 45° C. Iso-Propyl alcohol (150 mL) was added, charcaolized and filtered. The clear filtrate was acidified with 20%IPA HC1 to yield (1R, 2S)-Methoxamine HC1 which was filtered and dried (48 g); White crystalline powder; Purity by HPLC : 100%; Chiral purity : 100 %; mp : 172-175 °C; [α]D: -47.94° (c = 2% in MeOH)
Spectroscopic interpretation
The structure of the product, (1R,2S)-Methoxamine HC1 was confirmed with the help of the following spectroscopic data.
a) IR (cm1) (KBr)
O-H stretch at 3300, aromatic C-H stretch at 3065, aliphatic C-H stretch at 2938, HN-H str. at 2693 and 2580, benzenoid bands at 1609 and 1578, C-N stretch at 1277, C-O stretch at 1217, C-H out of plane bending of 1,2,4-tri- substituted benzene ring at 818.
b) 1H NMR (DMSO-d6 300 MHz) (δH)
0.91 (3H,d, -CH-CH3), 3.71 (3H, s, -OCH3), 3.75 (3H, s, -OCH3), 5.14 (1H, m, -CH- NH3+), 5.95 (1H, d, -CH-OH), 6.83-7.01 (3H, m, aromatic protons), 8.25 (3H, bs, HN-H) The -OH proton appears to have exchanged with the solvent.
c) 13C NMR (DMSO-d6, 300 MHz) (δC)
II. 44 (-CH-CH3), 49.22 (-OCH3), 55.24 (-OCH3), 55.70 (-CH-CH3), 66.49 (CH-OH),
III. 41 – 153.03 (aromatic carbons)
d) Mass spectrum (ESI, methanol)
[M+H]+ at m/z 212 (100), [M-H2O]+ at m/z 194 (15).
(IS, 2i?)-Methoxamine tartrate complex was added to water (275 mL) at 35 °C, basified
to 12 – 13 pH with 50% sodium hydroxide solution. Dichloromethane (250 mL) was added and stirred for 30 min. Separated the organic layer, dried over sodium sulphate and concentrated completely under vacuum at 45 °C. Iso-Propyl alcohol (175 mL) was added, charcaolized and filtered. The clear filtrate was acidified with 20%IPA HC1 to yield (1S, 2R)-Methoxamine HC1 which was filtered and dried (51 g) White crystalline powder; Purity by HPLC : 99.99%; Chiral purity . 100 %; mp . 172-175 °C;[α]D : + 47.9° (c = 2% in MeOH)
Spectroscopic interpretation
The structure of the product, (1S, 2R)-Methoxamine HC1 was confirmed with the help of the following spectroscopic data.
a) m (cm1) (KBr)
O-H stretch at 3265, aromatic C-H stretch at 3059, aliphatic C-H stretch at 2997, HN-H str. at 2658 and 2567, benzenoid bands at 1611 and 1587,
C-N stretch at 1294, C-O stretch at 1217, C-H out of plane bending of 1,2,4-tri- substituted benzene ring at 818.
b) 1H NMR (DMSO-d6,300 MHz) (δH)
0.91 (3H,d, -CH-CH3), 3.71 (3H, s, -OCH3), 3.75 (3H, s, -OCH3), 5.14 (1H, m, -CH- NH3+), 5.97 (1H, d, -CH-OH), 6.83-7.01 (3H, m, aromatic protons), 8.19 (3H, bs, HN-H) The -OH proton appears to have exchanged with the solvent.
c) 13C NMR (DMSO-d6,300 MHz) (δc)
II. 46 (-CH-CH3), 49.18 (-OCH3), 55.23 (-OCH3), 55.68 (-CH-CH3), 66.45 (CH-OH),
III. 42 – 153.02 (aromatic carbons)
d) Mass spectrum (ESI, methanol)
[M+H]+ at m/z 212 (100), [M-H2O]+ at m/z 194 (15).
Example 5
Preparation of dl-threo-methoxamine HC1 (11)
dl-erythro-methoxamine HC1 (120g; 0.48 mol) was dissolved in DM water (500 mL) at 30 – 35 °C and cooled to 10 – 15 °C. The clear solution was basified using 50 % sodium hydroxide solution and extracted in dichloromethane (250 mL). The organic layer was separated and concentrated under vacuum. The residue thus obtained was dissolved in toluene (200 mL) and was added slowly to acetic anhydride (120 g; 1.17mol) at 65 – 70 °C. The reaction mass was maintained under stirring and further cooled to 10 – 20 °C. Conc.Sulphuric acid (57.6g; 0.58mol) was added to the reaction mass slowly by maintaining the reaction mass at 10 – 200 C. The reaction mass was heated to 35 – 400 C for 3 hours and concentrated under vacuum at below 80 °C.
The reaction mass was cooled to 10 – 15 °C and was dissolved in DM water (250 mL). The mass was maintained for 3 h at reflux temperature and again cooled to 10 – 15 °C.
The pH was adjusted to 12 – 13 using 50% sodium hydroxide solution and extracted the d/-threo-Methoxamine base in dichloromethane (250 mL). Separated the organic layer and concentrated under vacuum. The concentrated mass was triturated with iso-Propyl alcohol (150 mL); acidified using 20% HC1 in iso-propyl alcohol. Distilled the iso- propyl alcohol completely to the final traces and acetone (300 mL) was added. The material precipitated, crude dl-threo-methoxamine HC1 was filtered. (85 g) Off white powder; Purity by HPLC: 99.4%; mp: 221-223 °C Spectroscopic interpretation
The structure of the product, di-threo-methoxamine HC1 was confirmed with the help of the following spectroscopic data.
a) IR (cm”1) (KBr)
O-H stretch at 3401, aromatic C-H stretch at 3005, aliphatic C-H stretch at 2924, HN-H str. at 2581 and 2490, benzenoid bands at 1609 and 1578, C-N stretch at 1277, C-0 stretch at 1215, C-H out of plane bending of 1,2,4-tri- substituted benzene ring at 802.
b) NMR (DMSO-d6,300 MHz) (δH)
1.2 (3H,d, -CH-CHs), 3.72 (3H, s, -OCH3), 3.75 (3H, s, -OCH3), 4.87 (1H, q, -CH-CH3),6.3 (1H, d, -CH-OH), 6.83-6.99 (3H, m, aromatic protons), 8.03 (3H, bs, HN-H) The -OH proton appears to have exchanged with the solvent.
c) 13C NMR (DMSO-d6, 300 MHz) (δC)
14.76 (-CH-CH3), 52.15 (-OCH3), 55.89 (-OCH3), 67.34 (CH-OH), 111.96 – 153.21 (aromatic carbons)
d) Mass spectrum (ESI, methanol)
[M+H]+ at m/z 212 (100), [M-H2O]+ at m/z 194 (52).
Example 6
Preparation of (1S,2S)- Methoxamine HC1 (6)
(IR, 2S)-Methoxamine HC1 (120 g; 0.48 mol) was dissolved in DM water (500 mL) at 30 -35 °C and cooled to 10 – 15 °C. The clear solution was basified using 50 % sodium hydroxide solution and extracted in dichloromethane (250 mL). The organic layer was separated and concentrated under vacuum. The residue thus obtained was dissolved in toluene (200 mL) and was added slowly to acetic anhydride (120 g; 1.17 mol) at 65 – 70 °C. The reaction mass was maintained under stirring and further cooled to 10 – 20 °C. Conc.sulphuric acid (57.6 g; 0.58 mol) was added to the reaction mass slowly by maintaining the reaction mass at 10 – 20 °C. The reaction mass was heated to 35 – 40 °C for 3 hours and concentrated under vacuum at below 80 °C.
The reaction mass was cooled to 10-15°C and was dissolved in DM water (250 mL). The mass was maintained for 3 h at reflux temperature and again cooled to 10 – 15 °C. The pH was adjusted to 12-13 using 50% sodium hydroxide solution and extracted the (1S, 2S)-Methoxamine base in dichloromethane (250 mL). Separated the organic layer and concentrated under vacuum The concentrated mass was triturated with iso-Propyl alcohol (150 mL); acidified using 20% HC1 in iso-propyl alcohol. Distilled the iso- propyl alcohol completely to the final traces and acetone (300 mL) was added. The material precipitated, crude (IS, 2S)-methoxamine HC1 was filtered. (86 g); White crystalline powder; Purity by HPLC . 99.8%; Chiral purity : 99.7%; mp : 172-175 °C; [α]D: + 30.739° (c = 2% in MeOH)
Spectroscopic interpretation
The structure of the product, (IS, 2S)-methoxamine HC1 was confirmed with the help of the following spectroscopic data.
a) IR (cm1) (KBr)
O-H stretch at 3356, aromatic C-H stretch at 3080, aliphatic C-H stretch at 2999, HN-H str. at 2641 and 2583, benzenoid bands at 1611 and 1506, C-N stretch at 1302, C-O stretch at 1229, C-H out of plane bending of 1,2,4-tri- substituted benzene ring at 812.
b) 1H NMR (DMSO-d6 300 MHz) (δH)
1.04 (3H,d, -CH-CH3), 3.72 (3H, s, -OCH3), 3.75 (3H, s, -OCH3), 4.90 (1H, m, -CH- CH3),6.07 (1H, d, -CH-OH), 6.84-7.01 (3H, d, aromatic protons), 8.15 (3H, bs, HN-H)
The -OH proton appears to have exchanged with the solvent.
c) 13C NMR (DMSO-d6, 300 MHz) (δC)
14.75 (-CH-CH3), 52.18 (-OCH3), 55.21 (-OCH3), 55.69 (-CH-CH3), 67.32 (CH-OH), 111.38 -153.01 (aromatic carbons)
d) Mass spectrum (ESI, methanol)
[M+H]+ at m/z 212 (100), [M-H2O]+ at m/z 194 (48).
Example 7
Preparation of (1R, 2R)-Methoxamine HC1 (5)
(IS, 2R)Methoxamine HC1 (120g; 0.48 mol) was dissolved in DM water (500 mL) at 30 – 35 °C and cooled to 10 – 15 °C. The clear solution was basified using 50 % sodium hydroxide solution and extracted in dichloromethane (250 mL). The organic layer was separated and concentrated under vacuum. The residue thus obtained was dissolved in toluene (200 mL) and was added slowly to acetic anhydride (120 g; 1.17mol) at 65 – 70 °C. The reaction mass was maintained under stirring and further cooled to 10 – 20 °C. Cone.Sulphuric acid (57.6g; 0.58mol) was added to the reaction mass slowly by maintaining the reaction mass at 10 – 20 °C. The reaction mass was heated to 35 – 40 °C for 3 hours and concentrated under vacuum at below 80 °C.
The reaction mass was cooled tol0-15°C and was dissolved in DM water (250 mL). The mass was maintained for 3 h at reflux temperature and again cooled to 10 – 15 °C. The pH was adjusted to 12-13 using 50% sodium hydroxide solution and extracted the (IR, 2i?)-Methoxamine base in dichloromethane (250 mL). Separated the organic layer and concentrated under vacuum. The concentrated mass was triturated with iso-Propyl alcohol (150 mL); acidified using 20% HC1 in iso-propyl alcohol Distilled the iso- propyl alcohol completely to the final traces and acetone (300 mL) was added. The material precipitated, crude (1R, 2R)-methoxamine HC1 was filtered. (90 g) White crystalline powder; Purity by HPLC: 99.1%, Chiral purity. 100%; mp: 172-175 °C;[α]D: -29.04° (c – 2% in MeOH)
Spectroscopic interpretation
The structure of the product, (1R, 2R)methoxamine HC1 was confirmed with the help of the following spectroscopic data.
a) IR (cm1) (KBr)
O-H stretch at 3356, aromatic C-H stretch at 3078, aliphatic C-H stretch at 2999, HN-H str. at 2619 and 2500, benzenoid bands at 1611 and 1508, C-N stretch at 1302, C-O stretch at 1229, C-H out of plane bending of 1,2,4-tri- substituted benzene ring at 812.
b) 1H NMR(DMSO-d6 300 MHz) (δH)
I. 04 (3H,d, -CH-CHa), 3.72 (3H, s, -OCH3), 3.75 (3H, s, -OCH3), 4.90 (1H, m, -CH- CH3),6.07 (1H, d, -CH-OH), 6.83-7.01 (3H, d, aromatic protons), 8.13 (3H, bs, HN-H) The -OH proton appears to have exchanged with the solvent.
c) 13C NMR (DMSO-d6 300 MHz) (δe)
II. 41 (-CH-CH3), 52.16 (-OCH3), 55.22 (-OCH3), 55.70 (-CH-CH3), 67.32 (CH-OH), III. 39-153.15 (aromatic carbons)
d) Mass spectrum (ESI, methanol)
[M+H]+ at m/z 212 (100), [M-H2O]+ at m/z 194 (44).
PATENT
http://www.google.com/patents/US8491931
(1,R,2S)-Methoxamine
To a stirred solution of methoxycarbonyl (MeOC) protected alcohol i.e. (1R,2S)-2-[(methoxycarbonyl)amino]-1-(2,5-dimethoxyphenyl)-1-propanol (4.0 g, 14.9 mmol) in methanol (175 cm3) was added a solution of KOH (4.06 g, 72.8 mmol in water (60 cm3). The solution was cooled and acidified with phosphoric acid (15% v/v). The solution was extracted with DCM (2×50 cm3) and the aqueous layer basified by the addition of K2CO3. The aqueous layer was extracted with diethyl ether (5×50 cm3) and the combined ethereal extracts dried (MgSO4) and concentrated under reduced pressure to give the product as a clear yellow oil (1.9 g, 61%), 1H NMR (250 MHz; C2HCl3) 0.84 (3H, d, J 7.0, CH3), 3.19-3.22 (1H, m, H-2), 3.71 (6H, s, 2×OCH3), 4.67 (1H, d, J 5.0, H-1), 6.66-6.72 (2H, m, ArH), 6.92 (1H, d, J 2.5, ArH).
(1R,2S)-Methoxamine hydrochloride
To an ice cooled solution of (1R,2S)-methoxamine (1.9 g, 9.00 mmol) in anhydrous diethyl ether (30 cm3) was passed a stream of dry HCl gas for 45 mins. The resultant precipitate was filtered by suction, washed with cold diethyl ether and dried under nitrogen to yield the title compound as a white solid. (1.5 g, 68%). 1H NMR (250 MHz; [C2H3]2SO) 0.89 (3H, d, J 6.8, CH3), 3.37-3.42 (1H,M,H-2), 3.71 (3H, s, OCH3), 3.75 (3H, s, OCH3), 5.12 (1H, s, H-1), 5.92 (1H, d, J 4.3, OH), 6.84 (1H, dd, J 8.8, 3.0, ArH), 6.92-7.00 (2H, m, ArH); HPLC.


//1R,2S-methoxamine
RACEMIC
 |
|
Title: Methoxamine
CAS Registry Number: 390-28-3
CAS Name: a-(1-Aminoethyl)-2,5-dimethoxybenzenemethanol
Additional Names: a-(1-aminoethyl)-2,5-dimethoxybenzyl alcohol; 2-amino-1-(2,5-dimethoxyphenyl)-1-propanol; b-hydroxy-b-(2,5-dimethoxyphenyl)isopropylamine; b-(2,5-dimethoxyphenyl)-b-hydroxyisopropylamine; 2,5-dimethoxynorephedrine
Molecular Formula: C11H17NO3
Molecular Weight: 211.26
Percent Composition: C 62.54%, H 8.11%, N 6.63%, O 22.72%
Literature References: a1-Adrenergic agonist. Prepn: Baltzly et al., US 2359707 (1944 to Burroughs Wellcome). Metabolism: A. Klutch, M. Bordun, J. Med. Chem. 10, 860 (1967). Clinical pharmacology: N. T. Smith, C. Whitcher, Anesthesiology 28, 735 (1967); P. D. Snashall et al., Clin. Sci. Mol. Med. 54, 283 (1978). HPLC determn in plasma: I. A. Al-Meshal et al., J. Liq. Chromatogr. 12, 1589 (1989). Therapeutic use: P. M. C. Wright et al., Anesth. Analg. 75, 56 (1992); L. Cabanes et al., N. Engl. J. Med. 326, 1661 (1992). Comprehensive description: A. M. Al-Obaid, M. M. El-Domiaty, Anal. Profiles Drug Subs. 20, 399-431 (1991).
Derivative Type: Hydrochloride
CAS Registry Number: 61-16-5
Trademarks: Vasoxine (Burroughs Wellcome); Vasoxyl (Burroughs Wellcome); Vasylox (Burroughs Wellcome)
Molecular Formula: C11H17NO3.HCl
Molecular Weight: 247.72
Percent Composition: C 53.33%, H 7.32%, N 5.65%, O 19.38%, Cl 14.31%
Properties: Crystals, mp 212-216°. pKa (25°C) 9.2. Very sol in water: One gram dissolves in 2.5 ml water, in 12 ml ethanol. Practically insol in ether, benzene, chloroform. pH of a 2% aq soln between 4.5 and 5.5.
Melting point: mp 212-216°
pKa: pKa (25°C) 9.2
Therap-Cat: Antihypotensive.
Keywords: a-Adrenergic Agonist; Antihypotensive.
|
Regulatory Approval Pathways: EU vs US

Regulatory Approval Pathways: EU vs US
 Drug Authorization Procedures in the EU
Sponsors have several options when seeking market approval for a new drug in Europe: a national authorization procedure, a decentralized procedure, a mutual recognition procedure and a centralized procedure. Depending on a product’s eligibility, each of these authorization routes offers different advantages and disadvantages to the sponsor, and these should be considered when setting up the market strategy of a product.
National Procedure
This procedure is used whenever a company wants to commercialize a product in only one EU Member State.
The National procedure is specific to each country. That is, each country within the EU has its own procedures for authorizing a marketing application for a new drug. Sponsors can find information regarding the requirements and procedure of each country on the websites of the regulatory agencies.
ADVANTAGES of National Procedure
There are some advantages in submitting…
View original post 3,049 more words
Enasidenib (AG-221)

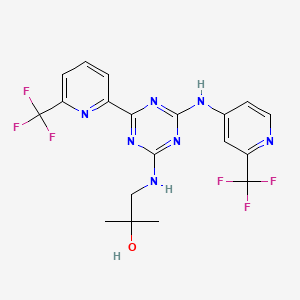
Enasidenib (AG-221)
1446502-11-9
Chemical Formula: C19H17F6N7O
Exact Mass: 473.13988
AG-221; AG 221; AG221; CC-90007; CC 90007; CC90007; Enasidenib
IUPAC/Chemical Name: 2-methyl-1-((4-(6-(trifluoromethyl)pyridin-2-yl)-6-((2-(trifluoromethyl)pyridin-4-yl)amino)-1,3,5-triazin-2-yl)amino)propan-2-ol
2-methyl-1-(4-(6-(trifluoromethyl)pyridin-2-yl)-6-(2-(trifluoromethyl)pyridin-4-ylamino)-1,3,5-triazin-2-ylamino)propan-2-ol
Agios Pharmaceuticals, Inc. innovator
Enasidenib, aslo known as AG-221 and CC-90007, is a potent and selective IDH2 inhibitor with potential anticancer activity (IDH2 = Isocitrate dehydrogenase 2). The mutations of IDH2 present in certain cancer cells result in a new ability of the enzyme to catalyze the NAPH-dependent reduction of α-ketoglutarate to R(-)-2-hydroxyglutarate (2HG). The production of 2HG is believed to contribute to the formation and progression of cancer . The inhibition of mutant IDH2 and its neoactivity is therefore a potential therapeutic treatment for cancer
AG-221 is an orally available, selective, potent inhibitor of the mutated IDH2 protein, making it a highly targeted investigational medicine for the potential treatment of patients with cancers that harbor an IDH2 mutation. AG-221 has received orphan drug and fast track designations from the U.S. FDA. In September 2013, Agios initiated a Phase 1 multicenter, open-label, dose escalation clinical trial of AG-221 designed to assess the safety and tolerability of AG-221 in advanced hematologic malignancies. In October 2014, Agios initiated four expansion cohorts as part of the ongoing Phase 1 study and expanded its development program with the initiation of a Phase 1/2 study of AG-221 in advanced solid tumors. For the detailed information of AG-221, the solubility of AG-221 in water, the solubility of AG-221 in DMSO, the solubility of AG-221 in PBS buffer, the animal experiment (test) of AG-221, the cell expriment (test) of AG-221, the in vivo, in vitro and clinical trial test of AG-221, the EC50, IC50,and affinity,of AG-221, For the detailed information of AG-221, the solubility of AG-221 in water, the solubility of AG-221 in DMSO, the solubility of AG-221 in PBS buffer, the animal experiment (test) of AG-221, the cell expriment (test) of AG-221, the in vivo, in vitro and clinical trial test of AG-221, the EC50, IC50,and affinity,of AG-221,
Agios Announces New Data from Ongoing Phase 1 Dose Escalation and Expansion Trial of AG-221 Showing Durable Clinical Activity in Patients with Advanced Hematologic Malignancies
IDH2-Mutant Inhibitor Shows Durable Responses of More than 15 Months in Patients with Advanced Acute Myeloid Leukemia (AML) and Other Blood Cancers
Proof-of-Concept Demonstrated in Myelodysplastic Syndrome (MDS) and Untreated AML
125-Patient Expansion Cohort and Global Registration-Enabling Program Remain on Track
Company to Host Conference Call and Webcast Today
CAMBRIDGE, Mass. & VIENNA–(BUSINESS WIRE)–Jun. 12, 2015– Agios Pharmaceuticals, Inc. (Nasdaq:AGIO), a leader in the fields of cancer metabolism and rare genetic disorders of metabolism, today announced new data from the dose-escalation phase and expansion cohorts from the ongoing Phase 1 study evaluating single agent AG-221, a first-in-class, oral, selective, potent inhibitor of mutant isocitrate dehydrogenase-2 (IDH2), in advanced hematologic malignancies. The data will be presented at the 20th Congress of the European Hematology Association (EHA) taking place June 11-14, 2015 in Vienna.
Data as of May 1, 2015 from 177 patients (104 in dose escalation and 73 from the first four expansion cohorts) with advanced hematologic malignancies treated with single agent AG-221 showed durable clinical activity and a favorable safety profile. More than half of the 177 patients remain on treatment. The study had an overall response rate of 40 percent (63 of 158 response-evaluable patients, using the criteria below) and a complete remission rate of 16 percent (26 of 158 response-evaluable patients). Patients responding to AG-221 continue to show durable clinical activity on treatment for more than 15 months, with an estimated 76 percent of responders staying on treatment for six months or longer. The overall safety profile observed was consistent with previously reported data with more than 100 additional patients treated as of the last analysis.
This new data reflects responses in the evaluable population, which includes all patients with a pre-AG-221 screening assessment and day 28 or later response assessment or an earlier discontinuation for any reason. Patients with a screening assessment who were still on treatment, but had not reached the day 28 disease assessment, were excluded.
“The clinical profile of AG-221 continues to be impressive from the perspectives of response rate, durability, safety and unique mechanism of action,” said Courtney DiNardo, M.D., lead investigator and assistant professor, leukemia atUniversity of Texas MD Anderson Cancer Center. “Additionally, it is encouraging to see early proof-of-concept in myelodysplastic syndrome (MDS) and untreated acute myeloid leukemia (AML) given the need for more effective therapies for these patients.”
“As the data from the AG-221 study continue to mature, we are compiling a robust dataset to quickly move this program into global registration studies later this year in collaboration with Celgene,” said Chris Bowden, M.D., chief medical officer of Agios. “We are excited about the speed of enrollment we’ve seen to date in our four expansion cohorts and are on track to enroll our recently announced fifth expansion cohort of 125 patients with relapsed and/or refractory AML. With this progress, we are executing on our strategy to combine speed and breadth to reach people with hematologic malignancies in urgent need of better treatments.”
About the Ongoing Phase 1 Trial for AG-221 in Advanced Hematologic Malignancies
AG-221 is currently being evaluated in an ongoing Phase 1 trial that includes a dose-escalation phase and four expansion cohorts of 25 patients each, evaluating patients with relapsed or refractory AML who are 60 years of age and older and transplant ineligible; relapsed or refractory AML patients under age 60; untreated AML patients who decline standard of care chemotherapy; and patients with other IDH2-mutant positive hematologic malignancies. Data reported here are from patients receiving AG-221 administered from 60 mg to 450 mg total daily doses in the dose escalation arm and 100 mg once daily in the first four expansion arms, as of May 1, 2015. The median age of these patients is 69 (ranging from 22-90). Treatment with AG-221 showed substantial reduction in the plasma levels of the oncometabolite 2-hydroxglutarate (2HG) to the level observed in healthy volunteers.
Safety Data
A safety analysis was conducted for all 177 treated patients as of May 1, 2015.
- The majority of adverse events reported by investigators were mild to moderate, with the most common being nausea, fatigue, increased blood bilirubin and diarrhea.
- The majority of serious adverse events (SAE) were disease related; SAEs possibly related to study drug were reported in 27 patients.
- A maximum tolerated dose (MTD) has not been reached.
- The all-cause 30-day mortality rate was 4.5 percent.
Efficacy Data
Sixty-three out of 158 response-evaluable patients achieved investigator-assessed objective responses for an overall response rate of 40 percent as of May 1, 2015.
- Of the 63 patients who achieved an objective response, there were 26 (16 percent) complete remissions (CR), three CRs with incomplete platelet recovery (CRp), 14 marrow CRs (mCR), two CRs with incomplete hematologic recovery (CRi) and 18 partial remissions (PR).
- Of the 111 patients with relapsed or refractory AML, 46 (41 percent) achieved an objective response, including 20 (18 percent) CRs, one CRp, 16 PRs, eight mCRs and one CRi.
- Of the 22 patients with AML that had not been treated, seven achieved an objective response, including three CRs, two PRs, one mCR and one CRi.
- Of the 14 patients with myelodysplastic syndrome (MDS), seven achieved an objective response, including two CRs, one CRp and four mCRs.
- Responses were durable, with duration on study drug more than 15 months and ongoing. As of the analysis date, an estimated 88 percent of responses lasted three months or longer, and 76 percent of responses lasted six months or longer.
Upcoming Milestones for AG-221
Agios studies in IDH2-mutated solid and hematologic tumors are ongoing or planned for 2015 to further support development of AG-221.
- Continue to enroll patients in the fifth expansion cohort of 125 patients with IDH2 mutant-positive AML who are in second or later relapse, refractory to second-line induction or re-induction treatment, or have relapsed after allogeneic transplantation.
- Initiate combination trials to evaluate AG-221 as a potential frontline treatment for patients with AML and a broad range of hematologic malignancies in the second half of 2015.
- Initiate a global Phase 3 registration-enabling study in relapsed/refractory AML patients that harbor an IDH2 mutation in the second half of 2015.
- Continue dose escalation in the Phase 1/2 trial in patients with advanced solid tumors, including glioma and angioimmunoblastic T-cell lymphoma (AITL) that carry an IDH2 mutation in 2015.
Conference Call Information
Agios will host a conference call and webcast from the congress to review the data on Friday, June 12, 2015, beginning at 8:00 a.m. ET (2:00 p.m. CEST). To participate in the conference call, please dial (877) 377-7098 (domestic) or (631) 291-4547 (international) and refer to conference ID 53010830. The webcast will be accessible live or in archived form under “Events & Presentations” in the Investors and Media section of the company’s website at www.agios.com.
About Agios/Celgene Collaboration
AG-221, the IDH1-mutant inhibitor AG-120 and the pan-IDH mutant inhibitor AG-881 are part of Agios’ global strategic collaboration with Celgene Corporation. Under the terms of the collaboration, Celgene has worldwide development and commercialization rights for AG-221. Agios continues to conduct clinical development activities within the AG-221 development program and is eligible to receive up to $120 million in payments on achievement of certain milestones and royalties on net sales. For AG-120, Agios retains U.S. development and commercialization rights. Celgene has an exclusive license outside the United States. Celgene is eligible to receive royalties on net sales in the U.S. Agios is eligible to receive royalties on net sales outside the U.S. and up to $120 million in payments on achievement of certain milestones. For AG-881, the companies have a joint worldwide development and 50/50 profit share collaboration, and Agios is eligible to receive regulatory milestone payments of up to $70 million.
About IDH Mutations and Cancer
IDH1 and IDH2 are two metabolic enzymes that are mutated in a wide range of hematologic and solid tumor malignancies, including AML. Normally, IDH enzymes help to break down nutrients and generate energy for cells. When mutated, IDH increases production of an oncometabolite 2-hydroxyglutarate (2HG) that alters the cells’ epigenetic programming, thereby promoting cancer. 2HG has been found to be elevated in several tumor types. Agios believes that inhibition of the mutated IDH proteins may lead to clinical benefit for the subset of cancer patients whose tumors carry them.
About Acute Myelogenous Leukemia (AML)
AML, a cancer of blood and bone marrow characterized by rapid disease progression, is the most common acute leukemia affecting adults. Undifferentiated blast cells proliferate in the bone marrow rather than mature into normal blood cells. AML incidence significantly increases with age, and according to the American Cancer Society, the median age of onset is 66. Less than 10 percent of U.S. AML patients are eligible for bone marrow transplant, and the vast majority of patients do not respond to chemotherapy and progress to relapsed/refractory AML. The five-year survival rate for AML is approximately 20 to 25 percent. IDH2 mutations are present in about 9 to 13 percent of AML cases.
About Myelodysplastic Syndrome (MDS)
MDS comprises a diverse group of bone marrow disorders in which immature blood cells in the bone marrow do not mature or become healthy blood cells. The National Cancer Institute estimates that more than 10,000 people are diagnosed with MDS in the United States each year. Failure of the bone marrow to produce mature healthy cells is a gradual process, and reduced blood cell and/or reduced platelet counts may be accompanied by the loss of the body’s ability to fight infections and control bleeding. For roughly 30 percent of the patients diagnosed with MDS, this bone marrow failure will progress to AML. Chemotherapy and supportive blood products are used to treat MDS.
About Agios Pharmaceuticals, Inc.
Agios Pharmaceuticals is focused on discovering and developing novel investigational medicines to treat cancer and rare genetic disorders of metabolism through scientific leadership in the field of cellular metabolism. In addition to an active research and discovery pipeline across both therapeutic areas, Agios has multiple first-in-class investigational medicines in clinical and/or preclinical development. All Agios programs focus on genetically identified patient populations, leveraging our knowledge of metabolism, biology and genomics. For more information, please visit the company’s website at agios.com.
clips
AG-221, Inhibitor Of IDH2 Mutants

The enzyme isocitrate dehydrogenase (IDH) is probably most famous for its role in the central cellular metabolic pathway, the Krebs cycle. The enzyme catalyzes the oxidative decarboxylation of isocitrate to α-ketoglutarate. One subtype of the enzyme, IDH1, is found in cells’ cytoplasm, and another, IDH2, is found in their mitochondria.

Target: IDH2
People with certain mutations in IDH end up making R-2-hydroxyglutarate (2-HG) instead of α-ketoglutarate. 2-HG is known to make cancer cells flourish. In fact, IDH mutations have been implicated in about 70% of brain cancers and have also been identified in solid tumors and blood cancers, such as acute myeloid leukemia.
Jeremy M. Travins of Agios Pharmaceuticals spoke about how scientists at the company found compounds based on substituted triazines that can cut down on 2-HG production by inhibiting a dimer of mutant IDH2. Using structure-activity relationships and a crystal structure of a lead compound bound to the mutant IDH2 dimer, they managed to develop a clinical candidate: AG-221. It turns out that AG-221 doesn’t bind to the active site of mutant IDH2. Rather, the compound binds to the spot where the two enzymes meet in the dimer.
Hitting this position in just the right way is tricky, Travins explained. Hydrogen-bonding interactions from the triazine and the two amino groups that flank it are critical.
The compound is in Phase I clinical trials, Travins said, and it’s been shown to lower 2-HG levels to those seen in people without cancer. What’s more, he noted, the drug candidate has few side effects, giving patients a higher quality of life than standard chemotherapeutic agents do.
Patent
http://www.google.com/patents/US20130190287
Compound 409—2-methyl-1-(4-(6-(trifluoromethyl)pyridin-2-yl)-6-(2-(trifluoromethyl)pyridin-4-ylamino)-1,3,5-triazin-2-ylamino)propan-2-ol

1H NMR (METHANOL-d4) δ 8.62-8.68 (m, 2H), 847-8.50 (m, 1H), 8.18-8.21 (m, 1H), 7.96-7.98 (m, 1H), 7.82-7.84 (m, 1H), 3.56-3.63 (d, J=28 Hz, 2H), 1.30 (s, 6H). LC-MS: m/z 474.3 (M+H)+.
| Patent ID | Date | Patent Title |
|---|---|---|
| US2013190287 | 2013-07-25 | THERAPEUTICALLY ACTIVE COMPOUNDS AND THEIR METHODS OF USE |
REFERENCES
1: Caino MC, Altieri DC. Molecular Pathways: Mitochondrial Reprogramming in Tumor Progression and Therapy. Clin Cancer Res. 2016 Feb 1;22(3):540-5. doi: 10.1158/1078-0432.CCR-15-0460. Epub 2015 Dec 9. PubMed PMID: 26660517; PubMed Central PMCID: PMC4738153.
2: Stein EM. IDH2 inhibition in AML: Finally progress? Best Pract Res Clin Haematol. 2015 Jun-Sep;28(2-3):112-5. doi: 10.1016/j.beha.2015.10.016. Epub 2015 Oct 19. Review. PubMed PMID: 26590767.
3: Rowe JM. Reasons for optimism in the therapy of acute leukemia. Best Pract Res Clin Haematol. 2015 Jun-Sep;28(2-3):69-72. doi: 10.1016/j.beha.2015.10.002. Epub 2015 Oct 22. Review. PubMed PMID: 26590761.
4: Stein EM. Molecular Pathways: IDH2 Mutations-Co-opting Cellular Metabolism for Malignant Transformation. Clin Cancer Res. 2016 Jan 1;22(1):16-9. doi: 10.1158/1078-0432.CCR-15-0362. Epub 2015 Nov 9. PubMed PMID: 26553750.
5: Kiyoi H. Overview: A New Era of Cancer Genome in Myeloid Malignancies. Oncology. 2015;89 Suppl 1:1-3. doi: 10.1159/000431054. Epub 2015 Nov 10. Review. PubMed PMID: 26551625.
6: Tomita A. [Progress in molecularly targeted therapies for acute myeloid leukemia]. Rinsho Ketsueki. 2015 Feb;56(2):130-8. doi: 10.11406/rinketsu.56.130. Japanese. PubMed PMID: 25765792.
/////////Enasidenib, AG-221,
CC(O)(C)CNC1=NC(C2=NC(C(F)(F)F)=CC=C2)=NC(NC3=CC(C(F)(F)F)=NC=C3)=N1
PF-06260414

PF-06260414
CAS: 1612755-71-1
Chemical Formula: C14H14N4O2S
Exact Mass: 302.0837
PF-06260414; PF 06260414; PF06260414; PF6260414; PF-6260414; PF 6260414.
IUPAC/Chemical Name: (R)-6-(4-methyl-1,1-dioxido-1,2,6-thiadiazinan-2-yl)isoquinoline-1-carbonitrile
- 6-[(4R)-4-Methyl-1,1-dioxido-1,2,6-thiadiazinan-2-yl]isoquinoline-1-carbonitrile
https://clinicaltrials.gov/ct2/show/NCT02070939
- 28 Jul 2015Discontinued – Phase-I for Cachexia in USA (PO)
- 27 Apr 2015Pfizer terminates a phase I trial (In volunteers) in USA (NCT02393807)
- 26 Mar 2015Pfizer plans a phase I pharmacokinetic trial for Healthy volunteers in USA (NCT02393807)
| Company | Pfizer Inc. |
| Description | Selective androgen receptor modulator |
| Molecular Target | Androgen receptor |
| Mechanism of Action | |
| Therapeutic Modality |
| Latest Stage of Development | Phase I |
| Standard Indication | Cachexia |
| Indication Details | Treat cachexia |
PF-06260414 is a selective androgen receptor modulator, or SARM, which is developed to treat muscle weakening. Testosterone’s anabolic properties help develop muscle mass, and its androgenic activity is associated with reproduction. Improving muscle mass would improve quality of life and may even prolong survival in certain patient populations.
PATENT
WO 2015173684
The androgen receptor (“AR”) is a ligand-activated transcriptional regulatory protein that mediates induction of male sexual development and function through its activity with endogenous androgens. Androgenic steroids play an important role in many physiologic processes, including the development and maintenance of male sexual characteristics such as muscle and bone mass, prostate growth,
spermatogenesis, and the male hair pattern. The endogenous steroidal androgens include testosterone and dihydrotestosterone (“DHT”). Steroidal ligands which bind the AR and act as androgens (e.g. testosterone enanthate) or as antiandrogens (e.g.
cyproterone acetate) have been known for many years and are used clinically.
6-[(4f?)-4-Methyl-1 , 1-dioxido-1 ,2,6-thiadiazinan-2-yl]isoquinoline-1-carbonitrile (Formula I), in its free base form, has the chemical formula C14H14N4SO2 and the following structural formula:

Formula I
Synthesis of 6-[(4f?)-4-methyl-1 , 1-dioxido-1 ,2,6-thiadiazinan-2-yl]isoquinoline-1-carbonitrile is disclosed in co-pending international patent application,
PCT/IB2013/060381 , filed 25th November 2013, and published as WO 2014/087298 on 12th June 2014, assigned to the assignee of the present invention and which is incorporated herein by reference in its entirety. 6-[(4f?)-4-Methyl-1 , 1-dioxido-1 ,2,6-thiadiazinan-2-yl]isoquinoline-1-carbonitrile is known to be active as a selective androgen receptor modulator (SARM) and, as such, is useful for treating and/or preventing a variety of hormone-related conditions, for example, conditions associated with androgen decline, such as, inter alia, anaemia; anorexia; arthritis; bone disease; musculoskeletal impairment; cachexia; frailty; age-related functional decline in the elderly; growth hormone deficiency; hematopoietic disorders; hormone replacement; loss of muscle strength and/or function; muscular dystrophies; muscle loss following surgery; muscular atrophy; neurodegenerative disease; neuromuscular disease;
obesity; osteoporosis; and, muscle wasting.
Identification of new solid forms of a known pharmaceutical active ingredient provide a means of optimising either the physicochemical, stability, manufacturability and/or bioperformance characteristics of the active pharmaceutical ingredient without modifying its chemical structure. Based on a chemical structure, one cannot predict with any degree of certainty whether a compound will crystallise, under what conditions it will crystallise, or the solid state structure of any of those crystalline forms. The specific solid form chosen for drug development can have dramatic influence on the properties of the drug product. The selection of a suitable solid form is partially dictated by yield, rate and quantity of the crystalline structure. In addition, hygroscopicity, stability, solubility and the process profile of the solid form such as compressibility, powder flow and density are important considerations.
The general reaction schemes provided herein illustrate the preparation of 6-[(4f?)-4-methyl-1 , 1-dioxido-1 ,2,6-thiadiazinan-2-yl]isoquinoline-1-carbonitrile (Formula I).

Example 1

Procedure:
Into a 2L 3-neck round bottom flask equipped with a mechanical stirrer, reflux condenser and thermocouple with heating mantle was placed 2-methyltetrahydrofuran (2-MeTHF) (10 mL/g; 8.15 moles; 817 ml_; 702 g) followed by racemic-2,2′-bis(diphenylphosphino)-1 ,1 ‘-binaphthyl (BINAP) (0.04 equiv (molar); 14.0 mmol; 8.74 g) and bis(dibenzylideneacetone)palladium (Pd2(dba)3) (0.04 equiv (molar); 14.0 mmol;
8.07 g). The mixture was degassed by pulling vacuum and refilling with nitrogen three times then heated to 75 °C for 15 minutes and cooled to ambient temperature. In a separate flask, (S)-3-amino-2-methylpropan-1-ol (1.60 equiv; 561 mmol; 50.0 g, prepared using literature methods, for example as disclosed in EP-A-0,089, 139 published on 21st September 1983) was dissolved in 2-methyltetrahydrofuran (5 ml_/g;
4.08 moles; 409 ml_; 351 g) and degassed by pulling vacuum and refilling with nitrogen three times. Into the pot containing the catalyst was added 6-(bromoisoquinoline-1- carbonitrile) (1.00 equiv; 351 mmol; 81.75 g) and cesium carbonate (1.6 equiv (molar); 561 mmol; 185 g) in single portions followed by the solution of the aminoalcohol via addition funnel. The reaction mixture was again degassed by pulling vacuum and refilling with nitrogen three times. The reaction was heated to 70 °C for 3 hours. The reaction was cooled to ambient temperature and filtered through a pad of Celite. The contents of the flask were rinsed out with three 100 mL portions of 2-methyltetrahydrofuran. The filtrate was transferred into a 2L round bottom flask equipped with a thermocouple and mechanical stirrer under nitrogen. Silica Gel (Silicylate SiliaMet® Thiol) (0.4 g/g-pure-LR; 544 mmol; 32.7 g) was charged and the flask was stirred at 40 °C overnight. The following morning, the reaction was cooled to < 30 °C and filtered again through Celite. The pad was washed with 100ml_ of 2-methyltetrahydrofuran (or until no yellow color persisted in the filtrate). The filtrate was placed into a 3L round bottom flask equipped with a magnetic stir bar, distillation head (with condenser and receiving flask), and thermocouple. The mixture was heated to 60 °C and placed under vacuum (-450-500 mbar) to distil out 1.3 L total of 2-methyltetrahydrofuran. 500 mL of toluene was added to precipitate the desired product. The heating mantle was removed and the reaction was allowed to reach ambient temperature. The mixture was stirred for 1 hour at ambient temperature and then the solids were collected by vacuum filtration on a sintered glass funnel. The cake was dried overnight on the funnel under vacuum. The following morning, the solids were transferred into an amber bottle and weighed (71.9 g; 298 mmol). The product was used in the next step without further purification.
Example 2

Procedure:
In a 1 L reactor equipped with a temperature probe and overhead stirring was added the product of Example 1 (20.0 g; 1.00 equiv; 82.9 mmol) and 2-methyltetrahydrofuran (2-MeTHF) (30 mL/g-pure-LR; 5.98 moles; 600 mL; 515 g). The reaction mixture was
gently warmed to 40°C to achieve partial solubility. The reaction was cooled to 0°C. Once the reaction reached 0°C methanesulfonyl chloride (MsCI) (1.4 equiv (molar); 1 16 mmol; 8.98 mL; 13.3 g) was added in a single portion followed immediately by triethylamine (TEA) (1.4 equiv (molar); 116 mmol; 16.2 mL; 11.7 g) dropwise via syringe over a period of 15 minutes. The reaction mixture was further stirred for 30 min at 0°C and then warmed to 23°C for 60 minutes. The product (26.47 g; 1.00 equiv; 82.88 mmol; 26.47 g; 100% assumed yield) was then used without purification for the sulfonylation reaction.
Example 3
t-BuOH, 2-MeTHF
o 0 °C to 23 °C o
CI-S-N=C=0 CI-S-NHBoc
0 O
Procedure:
To a solution of t-butyl alcohol (t-BuOH) (1 equiv (molar); 116 mmol; 1 1.0 mL; 8.60 g) in 2-methyltetrahydrofuran (2-MeTHF) (1 M; 1.16 moles; 116 mL; 99.6 g) at 0°C was added chlorosulfonyl isocyanate (116 mmol; 1.00 equiv; 10.1 mL; 16.4 g) dropwise. The homogeneous solution was stirred for 30 minutes at ambient temperature and then used directly in the sulfonylation reaction.
Example 4

Sulfonylation Reaction Procedure:
A previously prepared solution of the product of Example 3 (1.4 equiv (molar); 1 16 mmol; 116 g) in 2-methyltetrahydrofuran was added to a suspension of the product of Example 2 (1.00 equiv; 82.89 mmol; 26.5 g) at 0°C. The mixture was warmed to ambient temperature over 30 minutes. HPLC analysis revealed the reaction was complete. The reaction was quenched with a 10% sodium carbonate solution (2 equiv
(molar); 165 mmol; 101 mL; 1 17 g) and water (to dissolve salts) (5 L/kg; 7.35 moles; 132 mL; 132 g). The top organic layer was removed and passed through a plug of Carbon (Darco G60) (0.5 g/g) on a filter. A significant improvement in color (dark orange to yellow) was observed. The solution was concentrated to 10 total volumes and used in the next step without purification.
Example 5

Procedure:
A solution of the product of Example 4 (1.OOequiv; 82.9 mmol; 41.3 g) in 2-methyltetrahydrofuran (2-MeTHF) (10ml_/g; 4.12 moles; 413 mL; 355 g) was placed into a 1 L reactor equipped with an overhead stirrer and temperature probe. Next, potassium carbonate (K2CO3) (325 mesh) (6 equiv (molar); 497 mmol; 69.4 g) and water (0.0 L/100-g-bulk-LR; 459 mmol; 8.26 mL; 8.26 g) were added and the mixture heated to 40°C (jacket temperature) and stirred overnight. The reaction was cooled to ambient temperature and water (4L/kg-pure-LR; 9.17 moles; 165 mL; 165 g]) was added. The biphasic reaction was stirred for 1 hour at 23 °C. The aqueous layer was extracted and removed. The organic layer was passed through a plug of Carbon (Darco G60) (0.5 g/g-pure-LR; 20.7g) in a disposable filter. The 2-methyltetrahydrofuran solution was switched to a 10 volume solution of toluene via a constant strip-and-replace distillation to no more than 1 % 2-methyltetrahydrofuran. The toluene solution of the reaction product (1.00 equiv; 82.9 mmol; 33.4 g; 100% assumed yield) was used as-is in the next step without further purification.
Example 6

Procedure:
To a 1 L reactor under nitrogen and equipped with overhead stirring and a temperature probe was added the product of Example 5 (1.00 equiv; 78.7 mmol; 33.4 g) as a solution in toluene (10 mL/g-pure-LR; 3.00 moles; 317 ml_; 276 g). Next, trifluoroacetic acid (TFA) (10 equiv (molar); 787 mmol; 59.5 ml_; 89.8 g) was added to the reaction over a period of 1 hour keeping the internal temperature below 30°C. The dark red mixture was stirred for 1 hour. The reaction was quenched at 23 °C by the addition of sodium carbonate (5 equiv (molar); 394 mmol; 240 ml_; 278 g). The reaction was quenched slowly, over a period of 1 hour to form the TFA salt of the product. Once the charge was complete, the mixture was cooled to 0°C, held for 1 hour and filtered. The next morning, the solid product (6-[(4R)-4-methyl-1 , 1-dioxido-1 ,2,6-thiadiazinan-2-yl]isoquinoline-1-carbonitrile in its free base form) was weighed (0.89 equiv; 70.0 mmol; 21.2 g; 89.0% yield) and used in the next step without further purification.
Example 7
Crystalline 6-[(4f?)-4-methyl-1 , 1-dioxido-1 ,2,6-thiadiazinan-2-yl]isoquinoline-1-carbonitrile free base (Form (1)) was prepared as follows.
In a 1 L 3-neck round bottom flask was added 6-[(4R)-4-methyl-1 , 1-dioxido-1 ,2,6-thiadiazinan-2-yl]isoquinoline-1-carbonitrile free base (1.00 equiv; 70.0 mmol; 21.2 g) a magnetic stir bar and acetone (40ml_/g; 1 1.5 moles; 847 ml_; 669 g). The mixture was heated to reflux (approximately 57°C) and stirred for 1 hour. The mixture was concentrated by atmospheric distillation (heating mantle set at 65°C) and 40ml_ of acetone was collected into a graduated cylinder. Next, water (25 mL/g; 29.4 moles; 530 ml_; 530 g) was charged over a period of one hour. The mixture was stirred at ambient temperature for 60min before being cooled to 0°C at 1 °C /min for 1 hour. The solids were collected by filtration in a disposable funnel. Crystalline 6-[(4f?)-4-methyl-1 , 1-dioxido-1 ,2,6-thiadiazinan-2-yl]isoquinoline-1-carbonitrile (Form (1), 0.88 equiv; 61.9 mmol; 18.7 g; 88.3% yield) was dried under vacuum overnight at 40 °C. Typical purity after crystallization is 98%.
PATENT


Step 1. Synthesis of 6-bromoisoquinoline (#A1). A mixture of 4-bromobenzaldehyde (300.0 g, 1620.0 mmol) and amino acetaldehyde dimethyl acetal (170.4 g, 1620 mmol) in anhydrous toluene (1.5 L) was refluxed under a Dean-Stark condenser for 12 h. The solution was concentrated under vacuum. The residue was dissolved in anhydrous THF and cooled to —10° C. Ethyl chloroformate (193.3 mL, 1782 mmol) was added and stirred for 10 min at −10° C., and then allowed to warm to room temperature. Subsequently trimethyl phosphite (249.6 mL, 1782.0 mmol) was added dropwise to the reaction mixture and stirred for 10 h at room temperature. The solvent was evaporated under vacuum and the residue was dissolved in anhydrous DCM (1.5 L) and stirred for 30 minutes. The reaction mixture was cooled to 0° C., and titanium tetrachloride (1.2 L, 6480 mmol) was added dropwise. The reaction mixture was stirred at 40° C. for 6 days. The reaction mixture was poured into ice and pH was adjusted to 8-9 with aqueous 6N NaOH solution. The suspension was extracted three times with EtOAc. The organic layer was extracted with 3 M HCl. The acidic aqueous solution was adjusted to pH to 7-8 with 3N NaOH solutions and extracted two times with EtOAc. The organic layer was dried over anhydrous sodium sulfate and concentrated under reduced pressure to provide the product. Crude compound was dissolved in minimum amount of DCM and mixed with pentane to get compound #A1 as light brown solid. Yield: 90 g (35%). Rf: 0.6 (30% EtOAc in petroleum ether).
LCMS m/z=209 (M+1). 1H NMR (400 MHz, d6-DMSO): δ 7.82 (m, 2H), 8.11 (d, J=8.8 Hz, 2H), 8.30 (br s, 1H), 8.56 (d, J=6.0 Hz, 1H), 9.35 (s, 1H).
Step 2. Synthesis of 6-bromoisoquinoline 2-oxide (#A2). m-Chloroperoxybenzoic acid (120.0 g, 720.0 mmol) was added to a solution of #A1 (90.0 g, 480.0 mmol) in DCM (500 mL) at room temperature, and the reaction mixture was stirred for 16 h. 1N NaOH was added to the stirred reaction mixture to adjust the pH to 7-8. The layers were separated and the aqueous layer was extracted with DCM. The organic layer was dried over anhydrous sodium sulfate and concentrated under reduced pressure to render crude product. The solid product was triturated with the mixture of n-pentane and ethanol (8:2) to get the #A2 as white solid. Yield: 65 g (60%). Rf: 0.2 (EtOAc).
LCMS m/z=225 (M+1). 1H NMR (400 MHz, d6-DMSO): δ 7.83 (m, 2H), 7.91 (d, J=6.8 Hz, 1H), 8.21 (dd, J=8.0, 1.2 Hz, 1H), 8.26 (br s, 1H), 8.97 (s, 1H).
Step 3. Synthesis of 6-bromoisoquinoline-1-carbonitrile (#A3). Trimethylsilyl cyanide (52.0 mL, 580.0 mmol) was added dropwise to the stirred solution of #A2 (65.0 g, 290.0 mmol) and DBU (50.0 mL, 348.0 mmol) in THF (500 mL) at room temperature over a period of 15 minutes. The reaction mixture was stirred at room temperature for 1 h. Water was added to the reaction mixture, and the solution was extracted with DCM. The organic layer was dried over anhydrous Na2SO4 and concentrated under reduced pressure to give crude product. The product was purified by column chromatography using silica gel (100-200 mesh) with 0-4% EtOAc in petroleum ether as an eluent to give #A3 as white solid. Yield: 41 g (61%). Rf: 0.6 (30% EtOAc in petroleum ether).
LCMS m/z=233 (M+1). 1H NMR (400 MHz, d6-DMSO): δ 8.07 (dd, J=11.2, 2.0 Hz, 1H), 8.21 (m, 2H), 8.55 (br s, 1H), 8.77 (d, J=7.6 Hz, 1H).
A General Procedure to Prepare Intermediates of #A4, #A5, #A6 and #1, #2, #3, #4, #6, #7.
Step 4. A solution of #A3 (1 eq.) in toluene (50 mL) was degassed by bubbling with argon gas for 15 min and then Pd2dba3 (0.03 eq.), BINAP (0.06 eq.) and Cs2CO3(3 eq.) were added to the solution followed by the addition aminoalcohol (2 eq.). The mixture was heated at 100° C. under argon atmosphere for 3 h. Reaction mixture was cooled to room temperature, diluted with EtOAC and washed with water and brine. The organic layer was dried over Na2SO4 and concentrated to get crude product. The crude compounds were purified by silica gel (100-200 mesh) column chromatography by using 0-5% MeOH in DCM. Yields: 25-45%.
Step 5. MsCl (1 eq.) was added dropwise to a solution of #A4 (1 eq.) and Et3N (2 eq.) in DCM (10 mL) at 0° C. and was stirred at room temperature for 3 h. The reaction mixture was diluted with DCM, washed with water and brine. The organic layer was dried over Na2SO4 and concentrated. Crude products were used in next step without further purification.
Step 6. t-Butanol (2 eq.) was slowly added to a solution of chloro sulfonyl isocyanate (2 eq.) in toluene (1 mL/1 mmol) at 0° C. The reaction mixture was stirred at room temperature for 45 min. This solution (t-butyl chlorosulfonylcarbamate) was then added to a solution of #A5 (1 eq.) and DIPEA (4 eq.) in THF and stirred at room temperature for 12 h. Reaction mixture was diluted with water and extracted with EtOAc. Organic layer was washed with water, brine, then dried over anhydrous Na2SO4 and concentrated. Crude products were purified by silica gel (100-200 mesh) column chromatography using 0-40% EtOAc in petroleum ether.
Step 7. TFA was added to a solution of #A6 (1 eq.) in DCM (8 mL) at 0° C. and stirred at room temperature for 2 h. Reaction mixture was concentrated, diluted with water, neutralized with sat. aq. NaHCO3 soln. then extracted with DCM. The organic layer was washed with water and dried over Na2SO4 then concentrated. The crude products were purified by triturating with DCM and pentane to provide the compound. In the case of racemic materials, the enantiomers were separated by chiral preparative HPLC.
Column: CHIRALPAK IA, 4.6 mm×250, 5 μm; Mobile phase: n-Hexane: EtOH (65:35) (For X3: 35:65; For X2: 70:30); Flow rate: 1 mL/min; Eluent: EtOH.
EXAMPLE 16-[(3S)-3-methyl-1,1-dioxido-1,2,5-thiadiazolidin-2-yl]isocluinoline-1-carbonitrile (#1; R═CH3)
LCMS m/z=289.1 (M+1). 1H NMR (400 MHz, d6-DMSO): δ 1.37 (d, J=6.3 Hz, 3H), 3.27 (m, 1H), 3.74 (m, 1H), 4.63 (m, 1H), 7.17 (d, J=5.7 Hz, 1H), 7.72 (m, 1H), 7.89 (dd, J=10.7, 2.1 Hz, 1H), 8.26 (m, 2H), 8.62 (d, J=5.7 Hz, 1H).
PATENT
example 9
6 – [(3S) -3-methyl-1, 1 -dioxido-1, 2,5-thiadiazolidin-2-carbonitrile 1-yl1naphthalene
(Stereochemistry is arbitrarily Assigned)
LCMS m / z = 286.0 (M – H). 1 H NMR (400 MHz, cf 6 -DMSO): δ 1 .31 (d, J = 6.2 Hz, 3H), 3.13 – 3.25 (m, 1H), 3.71 (dt, J = 12.5, 6.8 Hz, 1H), 4.49 – 4.62 (m, 1H), 7.62 – 7.70 (m, 1H), 7.75 – 7.83 (m, 2H), 7.99 (t, J = 7.8 Hz, 1H), 8.07 (d, J = 6.6 Hz, 1H), 8.14 (d, J = 8.9 Hz, 1H), 8.28 (d, J = 8.4 Hz, 1H). Chiral HPLC purity: 99.1% (retention time 17.12 minutes)

Step 1. Synthesis of amino ester (# D1). Thionylchlride (8.5 mL, 1 16.5 mmol) Was added to the solution of amino acid (4.0 g, 38.8 mmol) in MeOH (170 mL) at 0 ° C, and the reaction mixture Was Stirred for 6 h at room temperature. The reaction Was monitored by TLC, and after-disappearance of the starting material It was cooled to room temperature and solid NaHC0 3 Was added. The reaction mixture Was filtered, concentrated in vacuo and the resulting and residue Was triturated with diethyl ether to crude obtenir # D1 (4 g, 90%) as a white solid. R f : 0.4 (f-BuOH: AcOH: H 2 0 (4: 0.5: 0.5)).
GCMS m / z 1 17.1 (M +). 1 H NMR (400 MHz, cf 6 -DMSO): δ 1.17 (d, J = 6.8Hz, 3H), 2.83 – 2.88 (m, 2H), 3.03 – 3.05 ( m, 1H), 3.65 (s, 3H), 8.02 – 8.30 (br s, 3H).
Step 2. Synthesis of aminoalcohol (# D2). # D1 (2.0 g, 13.0 mmol) Was added
portionwise to a suspension of LiAlH 4 (1.4 g, 39.2 mmol) in THF (75 mL) under nitrogen atmosphere at 0 ° C. The reaction mixture Was Stirred for 30 minutes and allowed to stir Then at room temperature for Reviews another 30 minutes. The reaction mixture Was Refluxed for 2 h, And Then It was cooled to -10 ° C and quenched with ice cold water Carefully (1.4 mL). 10% NaOH solution (2.8 mL) and ice cold water (4.2 mL) Were added, and the mixture Was Stirred for 15 minutes. It was filtered, and the filtrate washed with EtOAc (3 x 100 mL), dried over anhydrous Na 2 S0 4 and Concentrated under vacuum to obtenir # D2 (1.2 g, 86%) as a pale yellow liquid. R f: 0.2 (20% MeOH in DCM).
1 H NMR (400 MHz, cf 6 -DMSO): δ 0.78 (d, J = 6.8Hz, 3H), 1.46 – 1.54 (m, 1H), 2.41 -2.45 (m, 2H), 2.50 – 2.54 (m , 1H), 3.22 – 3.34 (m, 4H).
Step 3. Synthesis of coupling product (# D3). K 3 P0 4 (6.1 g, 28.8 mmol), BINAP (0.44 g, 0.72 mmol) and Pd 2 (dba) 3 (0.32.0 g, 0.36 mmol) Was added to the degassed
suspension of 6-bromo-1 -cyanoisoquinoline # A3 (1.7 g, 7.2 mmol), # D2 (1.2 g, 14.5 mmol) in DMSO at room temperature. The reaction mixture Was heated at 105 ° C for 2 h. The reaction Was cooled to room temperature, water (500 mL) Followed by EtOAc (100 mL) Were added, and the mixture Was Stirred for 10 minutes. The biphasic mixture Was filtered through a Celite ™ pad and washed with EtOAc (100 mL). The organic layer Was separated, and the aqueous layer Was Extracted with EtOAc (3 x 100 mL). The combined organic layers Were dried over anhydrous Na 2 S0 4 , concentrated under Reduced pressure to get a crude material. Reviews This was purified by column chromatography on 100-200 mesh silica gel, using 50-70% EtOAc in petroleum ether as the eluent to obtenir # D3 (0.5 g, 48.5%) as a yellow solid. R f : 0.4 (60% EtOAc in petroleum ether).
LCMS m / z = 242.0 (M + H). 1 H NMR (400 MHz, cf 6 -DMSO): δ 0.97 (d, J = 6.4Hz, 3H), 1.87 – 1.99 (m, 1H), 2.92 – 2.99 (m, 1H), 3.20 – 3.27 (m, 1H), 3.38 – 3.42 (m, 2H), 4.59 (t, J = 5.2 Hz, 1H), 6.77 (d, J = 2.0, 1H ), 7.01 (t, J = 5.6 Hz, 1H), 7.34 (dd, J = 9.2 Hz, J = 2.0 Hz, 1H), 7.73 (d, J = 6.0 Hz, 1H), 7.88 (d, J = 8.8 Hz, 1H), 8.312 (d, J = 6.0 Hz, 1H).
Step 4. Methanesulfonated coupling product (# D4). Triethylamine (0.44 mL, 3.1 mmol) Was added to a solution of # D3 (0.50 g, 2.0 mmol) in DCM at 0 ° C.
Methanesulfonylchloride (0.25 mL, 3.1 mmol) Was added over 10 minutes, and the reaction mixture Was Stirred for 1 h at room temperature. After disappearance of the starting material by TLC, It was diluted with DCM and washed with water. The organic layer Was separated, dried over Na 2 S0 4 , concentrated under pressure to obtenir Reduced crude # D4 (0.6 g, crude) as yellow solid. Reviews This was used for next step Without Any purification. R f : 0.6 (50% EtOAc in petroleum ether).
LCMS m / z = 320.0 (M + H). 1 H NMR (400 MHz, CDCl 3 ): δ 1.17 (d, J = 6.8Hz, 3H), 2.32 – 2.37 (m, 1H), 3.06 (s, 3H), 3.26 – 3.41 (m, 2H), 4.16 – 4.20 (m, 1H), 4.33 – 4.37 (m, 1H), 4.75 (br s, 1H), 6.70 (d, J = 2.4, 1 H), 7.09 (dd, J = 9.2 Hz, 2.4 Hz, 1H), 7.57 (d, J = 6.0 Hz, 1H), 8.05 (d, J = 9.2 Hz, 1H), 8.39 (d, J = 5.6 Hz, 1H).
Step 5. cyclized and uncyclized intermediates (# D5, D6 #). Chlorosulfonyl isocyanate (1.2 mL, 13.1 mmol) Was added dropwise to a solution of f-BuOH (1.4 mL, 13.1 mmol) in toluene (4.0 mL) at -5 ° C. The reaction mixture Was Stirred at room temperature for 20 minutes, And Then THF (1 mL) Was added to the resulting suspension to obtenir clear solution. In Reviews another flask, DIPEA (2.3 mL, 13.1 mmol) Was added to a solution of # D4 (0.6 g, 2.6 mmol crude) in dry THF (3 mL). The Above Prepared reagent (CIS0 2 NH-Soc) Was added to this reaction mixture dropwise at room temperature over a period of 20 minutes. The resulting and reaction mixture Was Then Stirred for 16 h at room temperature. The mixture Was diluted with EtOAc (100 mL) and washed with water (100 mL). The aqueous layer Was washed with EtOAc (2 x 100 mL), combined all the organic layers, dried over Na 2 S0 4 , concentrated under Reduced pressure to obtenir the crude product (LCMS shows Desired # D6 and uncyclized # D5. This crude Was purified by column chromatography on 100-200 mesh silica gel, using 10-30% EtOAc in petroleum ether as an eluent to obtenir Desired # D6 (0.35 g, 47.8%), and uncyclized # D5 (0.22 g, crude).
The uncyclized # D5 (0.22 g, crude) Was Dissolved in THF (1 mL) and DIPEA (0.6 ml) Was added to the solution. The reaction mixture Was Stirred Reviews another for 12 h at room temperature. After qui time, It was diluted with EtOAc (100 mL) and washed with water (100 mL). The aqueous layer Was washed with EtOAc (2 x 100 mL), combined all the organic layers, dried over Na 2 S0 4 , concentrated under pressure to obtenir Reduced crude product. Was this crude purified by column chromatography on 100-200 mesh silica gel, using 10-30% EtOAc in petroleum ether as an eluent to obtenir Desired # D6 (1 .1 g, 13.2%). Total amount of # D6 Was (0.5 g, 60% for two steps, 82% purity LCMS). R f : 0.8 (60% EtOAc in petroleum ether).
LCMS m / z = 403.1 (M + H). 1 H NMR (400 MHz, CDCl3): δ 1 .04 (d, J = 6.8 Hz, 3H), 1 .50 (s, 9H), 2.38 – 2.48 ( m, 1H), 3.65 – 3.82 (m, 2H), 3.92 – 4.02 (m, 1H), 4.30 – 4.38 (m, 1H), 7.79 – 7.81 (m, 1H), 7.86 – 7.88 (m , 2H), 8.34 – 8.37 (d, J = 9.2 Hz, 1H), 8.67 (d, J = 6.0 Hz, 1H).
Step 6. Racemate # D7 and final products (# 10, # 11). TFA (5 mL) Was added to a solution of # D6 (0.15 g, 0.37 mmol) in DCM (100 mL) at 0 ° C. The reaction mixture Was Stirred for 1 h at 0 ° C. The solution Was Neutralized with saturated aqueous NaHC03 solution at 0 ° C. The mixture Was diluted with water, Extracted with DCM (3 x 100 mL). The combined organic layers Were dried over anhydrous Na 2 S0 4 and Concentrated under pressure Reduced to obtenir racemic # D7 (0.10 mg, 73%).
LCMS m / z = 303.0 (M + H). R f : 0.3 (60% EtOAc in petroleum ether).
Enantiomeric separation: # D7 Was Submitted for chiral separation to obtenir final compounds # 10 (0.015 mg) and # 11 (0.016 mg).
Column: CHIRALPAK IA, 4.6 χ 250 mm, 5 m; Mobile phase: n-Hexane / / -PrOH / DCM (60% / 15% / 15%); Flow rate: 0.8 mL / min.
example 10
6 – [(4R) -4-methyl-1, 1 -dioxido-1, 2,6-thiadiazinan-2-yl1isoquinoline-1-carbonitrile (# 10; R = (R) -CH 3 )
LCMS m / z = 303.0 (M + 1). 1 H NMR (400 MHz, cf 6 -DMSO): δ 0.98 (d, J = 6.4Hz, 3H), 2.22 – 2.26 (m, 1H), 3.16 – 3.22 (m, 1H), 3.34 – 3.39 (m, 1H), 3.59 – 3.65 (m, 1H), 3.77 – 3.81 (m, 1H), 7.75 – 7.79 (m, 1H, Disappeared in D20 exchange), 7.95 (dd, J = 8.8 Hz, J = 2.0 Hz, 1H), 8.06 (d, J = 1 .6 Hz, 1H), 8.23 – 8.27 (m, 2H), 8703 (d, J = 5.2 Hz, 1H). R f : 0.3 (60% EtOAc in petroleum ether). Chiral HPLC purity: 98.2% (retention time on January 1 .43 minutes).
CLIP
PF-06260414, A Treatment For Muscle Diseases

Target: Androgen receptors
Disease: Muscular dystrophy, atrophy, sarcopenia

There aren’t many options when it comes to treating weakening muscles caused either by a disease such as muscular dystrophy or atrophy or by sarcopenia, the natural muscle weakening that comes with age. Doctors’ primary option is to give patients testosterone—a hormone with serious unwanted side effects on reproductive organs, the liver, and kidneys.


Pfizer’s Eugene Chekler spoke about PF-06260414, a selective androgen receptor modulator, or SARM, the company developed to treat muscle weakening. The idea, Chekler told C&EN, was to develop a nonsteroidal small molecule that would target androgen receptors but wouldn’t have any of testosterone’s negative side effects.

Testosterone’s anabolic properties help develop muscle mass, and its androgenic activity is associated with reproduction. To discover their SARM, Pfizer’s scientists used a novel screening strategy in which they decoupled anabolic and androgenic properties in vitro, Chekler said. Compounds that performed well in the muscle assay but had little effect in an assay that predicts androgenic response were developed further.
PF-06260414’s key pharmacophore is an isoquinoline with a pendant cyano group. The molecule also features a cyclic sulfuric diamide. It has completed Phase I clinical trials. “The market potential for this kind of treatment is huge,” Chekler said. “Improving muscle mass would improve quality of life and may even prolong survival in certain patient populations.”
Many answers from a first in human (FIH) study: Safety, tolerability, pharmacokinetics (PK) and pharmacodynamics (PD) of PF-06260414 in healthy Western and Japanese males
Annu Meet Am Soc Clin Pharmacol Ther (ASCPT) (March 8-12, San Diego) 2016, Abst PI-021
/////////////////////PF-06260414
N#CC1=NC=CC2=C1C=CC(N(C[C@H](C)CN3)S3(=O)=O)=C2
ND 0126

ND 0126
CAS 1240322-54-6
| Molecular Formula: | C29H25F3N6O3 |
|---|---|
| Molecular Weight: | 562.54241 g/mol |
methyl 5-[[2-methyl-5-[[3-(4-methylimidazol-1-yl)-5-(trifluoromethyl)benzoyl]amino]phenyl]methylamino]-1H-pyrrolo[2,3-b]pyridine-2-carboxylate
5-{2-Methyl-5-[3-(4-methyl-imidazol-1-yl)-5-trifluoromethyl-benzoylamino]-benzylamino}-1H-pyrrolo[2,3-b]pyridine-2-carboxylic Acid Methyl Ester
Potent dual ABL/SRC inhibitors based on a 7-azaindole core with the aim of developing compds. that demonstrate a wider activity on selected oncogenic kinases. Multi-Targeted Kinase Inhibitors (MTKIs) were then derived, focusing on kinases involved in both angiogenesis and tumorigenesis processes.
Dysfunction/deregulation of protein kinases (PK) is the cause of a large number of pathologies including oncological, immunological, neurological, metabolic and infectious diseases. This has generated considerable interest in the development of small molecules and biological kinase inhibitors for the treatment of these disorders.
Numerous PK are particularly deregulated during the process of tumorigenesis. Consequently protein kinases are attractive targets for anticancer drugs, including small molecule inhibitors that usually act to block the binding of ATP or substrate to the catalytic domain of the tyrosine kinase and monoclonal antibodies that specifically target receptor tyrosine kinases (RTK) and their ligands. In solid malignancies, it is unusual for a single kinase abnormality to be the sole cause of disease and it is unlikely that tumors are dependent on only one abnormally activated signaling pathway. Instead multiple signaling pathways are dysregulated. Furthermore, even single molecular abnormalities may have multiple downstream effects. Multi targeted therapy using a single molecule (MTKI = “Multi-Targeted Kinase Inhibitors”) which targets several signaling pathways simultaneously, is more effective than single targeted therapy. Single targeted therapies have shown activity for only a few indications and most solid tumors show deregulation of multiple signaling pathways. For example, the combination of a vascular endothelial growth factor receptor (VEGFR) inhibitor and platelet derived growth factor receptor (PDGFR) inhibitor results in a cumulative antitumor efficacy (Potapova et al, Mol Cancer Ther 5, 1280-1289, 2006).
Tumors are not built up solely of tumor cells. An important part consists of connective tissue or stroma, made up of stromal cells and extracellular matrix, which is produced by these cells. Examples of stromal cells are fibroblasts, endothelial cells and macrophages. Stromal cells also play an important role in the carcinogenesis, where they are characterized by upregulation or induction of growth factors and their receptors, adhesion molecules, cytokines, chemokines and proteolytic enzymes (Hofmeister et al., Immunotherapy 57, 1-17, 2007; Raman et al, Cancer Letters 256, 137-165, 2007; Fox et al, The Lancet Oncology 2, 278-289, 2001) The receptor associated tyrosine kinase VEGFR on endothelial and tumor cells play a central role in the promotion of cancer by their involvement in angiogenesis (Cebe-Suarez et al, Cell Mol Life Sci 63, 601-615, 2006). In addition, the growth factors TGF-β, PDGF and FGF2 secreted by cancer cells transform normal fibroblasts into tumor associated fibroblasts, which make their receptors a suitable target for inhibition by kinase inhibitors (Raman et al, 2007).
Moreover, increasing evidence suggests a link between the EGF receptor (EGFR) and HER2 pathways and VEGF-dependent angiogenesis and preclinical studies have shown both direct and indirect angiogenic effects of EGFR signaling (Pennell and Lynch, The Oncologist 14, 399-411, 2009). Upregulation of tumor pro -angiogenic factors and EGFR- independent tumor-induced angiogenesis have been suggested as a potential mechanism by which tumor cells might overcome EGFR inhibition. The major signaling pathways regulated by EGFR activation are the PI3K, MAPK and Stat pathways that lead to increased cell proliferation, angiogenesis, inhibition of apoptosis and cell cycle progression. EGFR is overexpressed in a wide variety of solid tumors, such as lung, breast, colorectal and cancers of the head and neck (Cook and Figg, CA Cancer J Clin 60, 222-243 2010). Furthermore, higher expression of EGFR has been shown to be associated with metastasis, decreased survival and poor prognosis.
c-Src, a membrane-associated non receptor tyrosine kinase, is involved in a number of important signal transduction pathways and has pleiotropic effects on cellular function. c-Src integrates and regulates signaling from multiple transmembrane receptor-associated tyrosine kinases, such as the EGFR, PDGFR, IGF1R, VEGFR, HER2. Together, these actions modulate cell survival, proliferation, differentiation, angiogenesis, cell motility, adhesion, and invasion (Brunton and Frame, Curr Opin Pharmacol 8, 427-432, 2008). Overexpression of the protein c-Src as well as the increase in its activity were observed in several types of cancers including colorectal, gastrointestinal (hepatic, pancreatic, gastric and oesophageal), breast, ovarian and lung (Yeatman, Nat Rev Cancer 4, 470-480, 2004).
The activation in EGFR or KRAS in cancers leads to a greatly enhanced level of Ras- dependent Raf activation. Hence, elimination of Raf function is predicted to be an effective treatment for the numerous cancers initiated with EGFR and KRAS lesions (Khazak et al, Expert Opin. Ther. Targets 11, 1587-1609, 2007). Besides activation of Raf signaling in tumors, a number of studies implicate the activation of the Ras-Raf-MAPK signaling pathway as a critical step in vasculo genesis and angiogenesis. Such activation is induced by growth factor receptors such as VEGFR2, FGFR2 and thus inhibition of Raf activation represents a legitimate target for modulation of tumor angiogenesis and vascularization.
Although VEGFR, PDGFR, EGFR, c-Src and Raf are important targets on both tumor cells and tumor stroma cells, other kinases such as FGFR only function in stromal cells and other oncogenes often only function in tumor cells.
Protein kinases are fundamental components of diverse signaling pathways, including immune cells. Their essential functions have made them effective therapeutic targets. Initially, the expectation was that a high degree of selectivity would be critical; however, with time, the use of “multikinase” inhibitors has expanded. Moreover, the spectrum of diseases in which kinase inhibitors are used has also expanded to include not only malignancies but also immune-mediated diseases / inflammatory diseases. The first step in signaling by multi-chain immune recognition receptors is mediated initially by Src family protein tyrosine kinases. MTKI targeting kinases involved in immune function are potential drugs for autoimmune diseases such as rheumatoid arthritis, psoriasis and inflammatory bowel diseases (Kontzias et al. , F 1000 Medicine Reports 4, 2012)
Protein kinases mentioned previously are also key components of many other physiological and pathological mechanisms such as neurodegeneration and neuroprotection (Chico et al, Nature Reviews Drug Discovery 8, 892-909, 2009), atherosclerosis, osteoporosis and bone resorption, macular degeneration, pathologic fibrosis, Cystogenesis (human autosomal dominant polycystic kidney disease…).
In WO2010/092489 and related patents/patent applications, we identified several compounds which exhibited interesting properties for such applications. However, we have discovered that some of these compounds could be enhanced in their properties by selectively working on particular regions of their structures. However, the mechanism of action of these structures on kinases was not precisely elucidated at the time of WO2010/092489’s filing and thus it was unexpectedly that we found the high activities of the structures disclosed in the present application. The subject matter of the present invention is to offer novel multi-targeted kinase inhibitors, having an original backbone, which can be used therapeutically in the treatment of pathologies associated with deregulation of protein kinases including tumorigenesis, human immune disorders, inflammatory diseases, thrombotic diseases, neurodegenerative diseases, bone diseases, macular degeneration, fibrosis, cystogenesis. The inhibitors of the present invention can be used in particular for the treatment of numerous cancers and more particularly in the case of liquid tumors such hematological cancers (leukemias) or solid tumors including but not limited to squamous cell cancer, small- cell lung cancer, non-small cell lung cancer, gastric cancer, pancreatic cancer, glial cell tumors such as glioblastoma and neurofibromatosis, cervical cancer, ovarian cancer, liver cancer, bladder cancer, breast cancer, melanoma, colorectal cancer, endometrial carcinoma, salivary gland carcinoma, renal cancer, prostate cancer, vulval cancer, thyroid cancer, sarcomas, astrocytomas, and various types of hyperproliferative diseases.

Efforts were made to improve a series of potent dual ABL/SRC inhibitors based on a 7-azaindole core with the aim of developing compounds that demonstrate a wider activity on selected oncogenic kinases. Multi-targeted kinase inhibitors (MTKIs) were then derived, focusing on kinases involved in both angiogenesis and tumorigenesis processes. Antiproliferative activity studies using different cellular models led to the discovery of a lead candidate (6z) that combined both antiangiogenic and antitumoral effects. The activity of 6z was assessed against a panel of kinases and cell lines including solid cancers and leukemia cell models to explore its potential therapeutic applications. With its potency and selectivity for oncogenic kinases, 6z was revealed to be a focused MTKI that should have a bright future in fighting a wide range of cancers.
5-{2-Methyl-5-[3-(4-methyl-imidazol-1-yl)-5-trifluoromethyl-benzoylamino]-benzylamino}-1H-pyrrolo[2,3-b]pyridine-2-carboxylic Acid Methyl Ester (6z)
Rational Design, Synthesis, and Biological Evaluation of 7-Azaindole Derivatives as Potent Focused Multi-Targeted Kinase Inhibitors
https://www.google.com/patents/WO2010092489A1?cl=en
Example 91: Preparation of methyl 5-(5-(3-(trifluoromethγl)-5~(4-methyl-1 H-imidazol-1 – yl)benzamido)-2-methγlbenzylamino)-1H-pyrrolo[2,3-blpyridine-2-carboχylate (ND0126)
Step 1 : preparation of methyl 5-(3-(trifluoromethyl)-5-(4-methyl-1 H-imidazol-1 – yl)benzamido)-2-methylbenzoate

The compound is obtained using the procedures of example 88 (step 4) replacing the 4-((3-(dimethylamino)pyrrolidin-1-yl)methyl)-3-(trifluoromethyl)-benzoic acid
(Shakespeare W. C, WO2007133562) by the 3-(trifluoromethyI)-5-(4-methyl-1H- imidazol-1-yl)benzoic acid.
Step 2: preparation of 3-(tπϊluoromethyl)-N-(3-formyl-4-methylphenyl)-5-(4- methyl-1H-imidazol-1-yl)benzamide

The compound is obtained by using the procedures of examples 83 (steps 1 and 2) replacing the methyl 5-(4-((4-methylpiperazin-1-yl)methyl)benzamido)-2- methylbenzoate with the methyl 5-(3-(trifluorometny))-5-(4-metbyl-1H-imidazol-1- yl)benzamido)-2-methylbenzoate.
Step 3: preparation of methyl 5-(5-(3-(trifluoromethyl)-5-(4-methyl-1 H-imidazol- 1-yl)benzamido)-2-methylbenzylamino)-1H-pyrrolo[2,3-bJpyridine-2-carboxylate (ND0126)
The composed is obtained according to example 83 (step 3) replacing N-(3-formyl-4- methylphenyl)-4-((4-methylpiperazin~1-yl)methyl)-benzamide with the 3- (trifluoromethyl)-N-(3-formyl-4-methylphenyl)-5-(4-methyl-1 H-imidazol-1-yl)benzamide.
PATENT
REFERENCES
| WO2005063747A1 * | Dec 23, 2004 | Jul 14, 2005 | Pfizer Italia S.R.L. | PYRROLO[2,3-b] PYRIDINE DERIVATIVES ACTIVE AS KINASE INHIBITORS, PROCESS FOR THEIR PREPARATION AND PHARMACEUTICAL COMPOSITION COMPRISING THEM |
| WO2008028617A1 * | Sep 4, 2007 | Mar 13, 2008 | F. Hoffmann-La Roche Ag | Heteroaryl derivatives as protein kinase inhibitors |
| WO2008124849A2 * | Apr 10, 2008 | Oct 16, 2008 | Sgx Pharmaceuticals, Inc. | Pyrrolo-pyridine kinase modulators |
| WO2008144253A1 * | May 9, 2008 | Nov 27, 2008 | Irm Llc | Protein kinase inhibitors and methods for using thereof |
| WO2014102376A1 * | Dec 30, 2013 | Jul 3, 2014 | Oribase Pharma | Protein kinase inhibitors |
| WO2014102377A1 * | Dec 30, 2013 | Jul 3, 2014 | Oribase Pharma | Azaindole derivatives as multi kinase inhibitors |
| WO2014102378A1 * | Dec 30, 2013 | Jul 3, 2014 | Oribase Pharma | Azaindole derivatives as inhibitors of protein kinases |
| US20150353540 * | Dec 30, 2013 | Dec 10, 2015 | Oribase Pharma | Azaindole derivatives as inhibitors of protein kinases |
| US2011312959 | 2011-12-22 | Derivatives of Azaindoles as Inhibitors of Protein Kinases ABL and SRC |
///////ND 0126, 1240322-54-6, PRECLINICAL
O=C(OC)c1cc2cc(cnc2n1)NCc3cc(ccc3C)NC(=O)c4cc(cc(c4)n5cc(C)nc5)C(F)(F)F
CC1=C(C=C(C=C1)NC(=O)C2=CC(=CC(=C2)N3C=C(N=C3)C)C(F)(F)F)CNC4=CN=C5C(=C4)C=C(N5)C(=O)OC


