Home » cancer
Category Archives: cancer
 DRUG APPROVALS BY DR ANTHONY MELVIN CRASTO
.....FOR BLOG HOME CLICK HERE
Flag and hits
ORGANIC SPECTROSCOPY

SUBSCRIBE

 Subscribe in a reader
Enter your email address:
Delivered by FeedBurner
Subscribe to New Drug Approvals by Email


Recent Comments
 | shivkr2 on SPSR Excellence Award 2025 – S… |
 | Bonkasaurus on Infigratinib phosphate |
 | PALOVAROTENE | ORGAN… on Palovarotene |
| Pfizer just purchase… on Temanogrel | |
| Pfizer To Cash In On… on Temanogrel |
Iberdomide
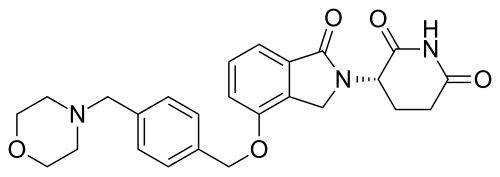
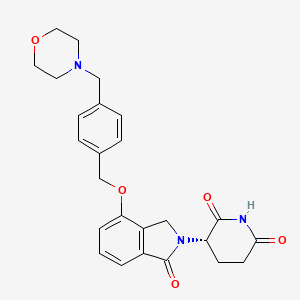
Iberdomide
CAS 1323403-33-3
as HCl: 1560678-63-8
MW 449.5 g/mol, C25H27N3O5
(S)-3-(4-((4-(Morpholinomethyl)benzyl)oxy)-1-oxoisoindolin-2-yl)piperidine-2,6-dione
(3S)-3-[7-[[4-(morpholin-4-ylmethyl)phenyl]methoxy]-3-oxo-1H-isoindol-2-yl]piperidine-2,6-dione
8/13/2026, APPROVAL 2026, FDA 2026, Zenbexus, cc-220, cc 220, 8V66F27X44, 79L3645KFI
To be used in combination with daratumumab and hyaluronidase-fihj and dexamethasone for adults with multiple myeloma who have received at least one prior line of therapy, including a proteasome inhibitor and an immunomodulatory agent
Iberdomide is a modulator of the E3 ubiquitin ligase complex containing cereblon (CRL4-CRBN E3 ubiquitin ligase), with immunomodulating and pro-apoptotic activities. Upon administration, iberdomide specifically binds to the cereblon (CRBN) part of the ligase complex, thereby affecting the ubiquitin E3 ligase activity, and targeting certain substrate proteins for ubiquitination. This induces the proteasome-mediated degradation of certain transcription factors, including Ikaros (IKZF1) and Aiolos (IKZF3) which are transcriptional repressors in T-cells. This leads to a reduction of their protein levels, and the modulation of the immune system, including activation of T-lymphocytes. In addition, this leads to a downregulation of other proteins, including interferon regulatory factor 4 (IRF4), which plays a key role in the proliferation of certain cancer cell types. CRBN, the substrate recognition component of the E3 ubiquitin ligase complex, plays a key role in the ubiquitination of certain proteins.
Iberdomide, sold under the brand name Zenbexus, is an anti-cancer medication used for the treatment of multiple myeloma.[1] It is a cereblon-modulating protein degrader[1] and a thalidomide analog.[2]. It is taken By mouth.[1]
Iberdomide was approved for medical use in the United States in August 2026.[3]
Medical uses
Iberdomide is indicated in combination with daratumumab, hyaluronidase, and dexamethasone for the treatment of adults with multiple myeloma who have received at least one prior line of therapy including a proteasome inhibitor and an immunomodulatory agent.[3]
Society and culture
Legal status
Iberdomide was approved for medical use in the United States in August 2026.[12] The U.S. Food and Drug Administration (FDA) granted the application for iberdomide priority review, breakthrough therapy, and orphan drug designations.[3]
Names
Iberdomide is the international nonproprietary name.[13]
Iberdomide is sold under the brand name Zenbexus.[14]
SYN
SYN

PAT
https://patentscope.wipo.int/search/en/detail.jsf?docId=WO2011100380&_cid=P12-MSZHNS-67589-1
5.2 3-[4-(4-MORPHOLIN-4-YLMETHYL-BENZYLOXY)-1-OXO- 1,3-DIHYDRO-ISOINDOL-2-YL]-PIPERIDINE-2,6-DIONE

Step 3 : To the THF solution of methyl 5-amino-4-(4-(4- (morpholinomethyl)benzyloxy)-1-oxoisoindolin-2-yl)-5-oxopentanoate (40 g, 83 mmol), was added potassium 2-methylpropan-2-olate (9.80 g, 87 mmol) portion wise at 0°C. The mixture was stirred at this temperature for 30 minutes. To the reaction mixture, was added 45 mL of 1N HCl solution, followed by 200 mL of saturated NaHCO3 solution. The mixture was diluted with 500 mL of EtOAc at 0°C, stirred for 5 minutes and separated. The organic layer was washed with water (50 mL × 3) and brine (100 mL), and concentrated on rota-vap to give a white solid, which was stirred in diethyl ether (300 mL) to give a suspension. The suspension was filtered to give 3-[4-(4-morpholin-4-ylmethyl-benzyloxy)-1-oxo-1,3-dihydro-isoindol-2-yl]-piperidine-2,6-dione as white solid (28.5g, 72% yield): HPLC: Waters Symmetry C18, 5μm, 3.9 × 150 mm, 1 mL/min, 240 nm, gradient to 95/5 acetonitrile/0.1% H3PO4 in 5 min,: tR = 4.78 min (98.5%); mp: 209-21 1 °C; 1H NMR (DMSO-d6) δ 1.86 – 2.09 (m, 1H, CHH), 2.29 – 2.38 (m, 4H, CH2,CH2), 2.44 (dd, J = 4.3, 13.0 Hz, 1H, CHH), 2.53 – 2.64 (m, 1H, CHH), 2.82 – 2.99 (m, 1H, CHH), 3.46 (s, 2H, CH2), 3.52 – 3.61 (m, 4H, CH2,CH2), 4.18 – 4.51 (m, 2H, CH2), 5.11 (dd, J = 5.0, 13.3 Hz, 1H, NCH), 5.22 (s, 2H, CH2), 7.27 – 7.38 (m, 5H, Ar), 7.40 – 7.53 (m, 3H, Ar), 10.98 (s, 1H, NH); 13C NMR (DMSO-d6) δ 22.36, 31.21, 45.09, 51.58, 53.14, 62.10, 66.17, 69.41,
114.97, 115.23, 127.64, 128.99, 129.81, 129.95, 133.31, 135.29, 137.68, 153.50, 168.01,
170.98, 172.83; LCMS: 465; Anal Calcd for C25H27N3O5 + 0.86 H2O: C, 64.63; H, 6.22; N,
9.04; Found: C, 64.39; H, 6.11; N, 8.89; H2O, 3.24.
5.61 (S)-3-[4-(4-MORPHOLIN-4-YLMETHYL-BENZYLOXY)-1-OXO-1,3- DIHYDRO-ISOINDOL-2-YL]-PIPERIDINE-2,6-DIONE

[386] Step 1 : Preparation of (S)-4-[4-(4-Bromomethyl-benzyloxy)-1-oxo-1,3-dihydro-isoindol-2-yl]-4-carbamoyl-butyric acid methyl ester
To a 2-L round bottom flask was charged methyl 5-amino-4-(4-hydroxy-1-oxoisoindolin-2-yl)-5-oxopentanoate (30 g, 103 mmol), 1,4-bis(bromomethyl)benzene (81 g, 308 mmol) and potassium carbonate (14.19 g, 103 mmol) and acetonitrile (1.2 L). The mixture was stirred at room temperature for 10 min and heated to 50°C for 12 hours. The reaction mixture was allowed to cool to room temperature. The mixture was filtered and the filtrate was concentrated on rota-vap. The resulted solid was dissolved in CH2Cl2 and loaded on 2 silica gel columns (330 g each) eluted using CH2Cl2/MeOH to give 4-[4-(4-bromomethyl-benzyloxy)-1-oxo-1,3-dihydro-isoindol-2-yl]-4-carbamoyl-butyric acid methyl ester as white solid (40g, 82%). 1H NMR (DMSO-d6) δ 1.98 – 2.13 (m, 1H, CHH), 2.14 – 2.23 (m, 1H, CHH), 2.23 – 2.32 (m, 2H, CHH, CHH), 3.50 (s, 3H, CH3), 4.34 – 4.63 (m, 2H, CH2), 4.67 – 4.80 (m, 3H, CH2, NCH), 5.25 (s, 4H, CH2), 7.19 (s, 1H, NHH), 7.24 – 7.34 (m, 2H, Ar), 7.41 – 7.54 (m, 5H, Ar), 7.58 (br. s., 1H, NHH)
[387] Step 2: Preparation of (S)-4-Carbamoyl-4-[4-(4-morpholin-4-ylmethyl-benzyloxy)-1-oxo-1,3-dihydro-isoindol-2-yl]-butyric acid methyl ester
To the CH2Cl2 solution of methyl 5-amino-4-(4-(4-(bromomethyl)benzyloxy)-1-oxoisoindolin-2-yl)-5-oxopentanoate (36.5 g, 77 mmol) was added morpholine (14.72 ml, 169 mmol) at 0 °C. The mixture was stirred at 0 °C for 1 hr. The mixture was added 200 mL of CH2Cl2, washed with water (100mL × 2) and brine (100 ml), dried in Na2SO4 and concentrated to give (S)-4-Carbamoyl-4-[4-(4-morpholin-4-ylmethyl-benzyloxy)-1-oxo-1,3-dihydro-isoindol-2-yl]-butyric acid methyl ester as white foam (39 g, 100%). M.p. 66-68 °C; Waters Symmetry C-18, 3.9 X 150 mm, 5 micro, 1 mL/min, 240 nm, isocratic 15/85 CH3CN/ 0.1% H3PO4 in H2O: 7.92 min (99%). 1H NMR (DMSO-d6) δ 2.00 – 2.12 (m, 1HH CHH), 2.14 – 2.22 (m, 1H, CHH), 2.22 – 2.29 (m, 2H, CHH,CHH), 2.30 – 2.39 (m, 4H, CH2,CH2), 3.46 (s, 2H, CH2), 3.50 (s, 3H, CH3), 3.53 – 3.63 (m, 4H, CH2,CH2), 4.28 – 4.59 (m, 2H, CH2), 4.73 (dd, J= 4.7, 10.2 Hz, 1H, NCH), 5.22 (s, 2H, CH2), 7.14 – 7.23 (m, 1H, NHH), 7.26 – 7.39 (m, 4H, Ar), 7.41 – 7.51 (m, 3H, Ar), 7.58 (s, 1H, NHH). 13C NMR (DMSO-d6) δ 24.82, 30.33, 44.78, 51.24, 53.12, 53.38, 62.09, 66.14, 69.35, 114.66, 115.12, 127.60, 129.00, 129.55, 130.18, 133.43, 135.31, 137.66, 153.42, 167.84, 171.73, 172.46; Anal Calcd for C26H31N3O6+ 0.3 H2O: C% 64.13; H% 6.54; N% 8.63; Found: C% 63.89; H% 6.39; N% 8.56.
[388] Step 3: Preparation of (S)-3-[4-(4-morpholin-4-ylmethyl-benzyloxy)-1-oxo- 1 , 3-dihydro-isoindol-2-yl]-piperidine-2,6-dione
To the THF solution of (S)-methyl 5-amino-4-(4-(4-(morpholinomethyl)benzyloxy)-1-oxoisoindolin-2-yl)-5-oxopentanoate (45 g, 93 mmol) was added potassium 2-methylpropan-2-olate (10.49 g, 93 mmol) portion wise (2g X5) at -78 °C. The mixture was stirred at this temperature for 30 min then was added 250 mL of 1N HCl solution followed by 200 mL of saturated NaHCO3 solution. The mixture was extracted with CH2Cl2 (150 mLx2). The organic layer was washed with water (50 mL × 3) and brine (100 mL), concentrated on rota-vap to give a white solid, which was then recrystallized from CH3CN
(100 mL) to give (S)-3-[4-(4-morpholin-4-ylmethyl-benzyloxy)-1-oxo-1,3-dihydro-isoindol-2-yl]-piperidine-2,6-dione as white solid (32g, 76%). mp: 140-142 °C. LC-MS m/e= 450. HPLC: Waters Symmetry C18, 5μm, 3.9 x 150 mm, 1 mL/min, 240 nm, isocratic
15/85 CH3CN/0.1% H3PO4 in 5 min,: tR = 5.61 min (99.5%); Chiral AGP C 18 4.0 × 150 mm, 5 μm 10/90 i-propanol/ 10 mM NH4Ac in 20 min,: tR = 10.07 min (99.5%); 1H NMR
(DMSO-d6) δ 2.28 – 2.38 (m, 4H, CH2,CH2), 2.44 (dd, J= 4.2, 13.1 Hz, 1H, CHH), 2.53- 2.63 (m, 1H, CHH), 2.79 – 3 02 (m, 1H, CHH), 3.49 – 3.69 (m, 4H, CH2,CH2), 4.11 – 4.52
(m, 2H, CH2), 5.11 (dd, J= 5.1, 13.2 Hz, 1H, NCH), 5.22 (s, 2H, CH2), 7.33 (d, J= 7.7 Hz,
4H, Ar), 7.40 – 7.52 (m, 3H, Ar), 10.97 (s, ΙΗ, ΝΗ). 13C NMR (DMSO-d6) δ 22.33, 31.18,
45.06, 51.55, 53.11, 62.07, 66.14, 69.38, 114.96, 115.20, 127.61, 128.97, 129.78, 129.93,
133.28, 135.27, 137.67, 153.48, 167.97, 170.95, 172.80. LC-MS: 465; Anal Calcd for
C25H27N3O5 C: 66.80%; H: 6.05%; N: 9.35%. Found: C:66.59%; H:5.79%; N:9.26%.
PAT
- PRMT5 inhibitors and uses thereofPublication Number:US-12448388-B2Grant Date:2025-10-21
- KRAS G12D modulating compoundsPublication Number:US-12448400-B2Grant Date:2025-10-21
- Arylmethoxy isoindoline derivatives and compositions comprising them and methods of use thereofPublication Number:ES-2956743-T3Priority Date:2010-02-11Grant Date:2023-12-27
- Arylmethoxy isoindoline derivatives and compositions comprising and methods of using the samePublication Number:AU-2013245487-A1Priority Date:2010-02-11
- Arylmethoxy isoindoline derivatives and compositions comprising and methods of using the samePublication Number:AU-2013245487-B2Priority Date:2010-02-11Grant Date:2016-03-10
- Arylmethoxy isoindoline derivatives and compositions comprising and methods of using the samePublication Number:EP-3599236-B1Priority Date:2010-02-11Grant Date:2023-08-23
- Treating cancerPublication Number:US-2025325586-A1
- Arylmethoxy isoindoline derivatives and compositions comprising and methods of using the samePublication Number:US-9822094-B2Priority Date:2010-02-11Grant Date:2017-11-21
- Arylmethoxy isoindoline derivatives and compositions comprising and methods of using the samePublication Number:EP-4289838-A2Priority Date:2010-02-11
- Arylmethoxy isoindoline derivatives and compositions comprising and methods of using the samePublication Number:US-2018037567-A1Priority Date:2010-02-11
- Arylmethoxy isoindoline derivatives and compositions comprising them and methods of using themPublication Number:ES-2713482-T3Priority Date:2010-02-11Grant Date:2019-05-22
- Arylmethoxy Isoindoline Derivatives and Compositions Comprising and Methods of Using the SamePublication Number:US-2011196150-A1Priority Date:2010-02-11
ADVERISEMENT
ANAX LABORATORIES
WEBSITE https://www.anaxlab.com/
Discovery Solutions, Supporting the chemistry needs of clients in the Medical, Analytical and Bio Sciences
Development Solutions, Developing from Lab scale to PR&D, Kilo Scale-ups and Commercial Scales
SEE MORE………Integrated Solutions, Manufacturing Solutions, Products,
Can’t Find? Let’s Connect
Phone : +91 897704 2010 / +91 9177075735, Email : info@anaxlab.com
#MedicinalChemistry, #DrugDiscovery, #OrganicSynthesis, #ChemicalLibrary, #BuildingBlocks, #SARStudies, #ChemistryInnovation, #medchem, #Drugdevelopment, #Biotech, #Biotechnology, #AnaxLaboratories, #Pharma


AS ON FEB2026 4.574 LAKHS VIEWS ON BLOG WORLDREACH AVAILABLEFOR YOUR ADVERTISEMENT
join me on Linkedin
Anthony Melvin Crasto Ph.D – India | LinkedIn
join me on Researchgate
RESEARCHGATE

join me on Facebook
Anthony Melvin Crasto Dr. | Facebook
join me on twitter
Anthony Melvin Crasto Dr. | twitter
+919321316780 call whatsaapp
EMAIL. amcrasto@gmail.com

References
References
- “U.S. Prescribing Information” (PDF). Packageinserts.bms.com. Retrieved 18 August 2026.
- Gao, Shaobing; Wang, Shichao; Song, Yongping (December 2020). “Novel immunomodulatory drugs and neo-substrates”. Biomarker Research. 8 (1): 2. doi:10.1186/s40364-020-0182-y. PMC 6953231. PMID 31938543.
- “FDA grants accelerated approval to iberdomide with daratumumab and hyaluronidase-fihj and dexamethasone for multiple myeloma”. U.S. Food and Drug Administration (FDA). 13 August 2026. Retrieved 16 August 2026.
 This article incorporates text from this source, which is in the public domain.
- Ye, Ying; Gaudy, Allison; Schafer, Peter; Thomas, Michael; Weiss, Daniel; Chen, Nianhang; et al. (May 2021). “First-in-Human, Single- and Multiple-Ascending-Dose Studies in Healthy Subjects to Assess Pharmacokinetics, Pharmacodynamics, and Safety/Tolerability of Iberdomide, a Novel Cereblon E3 Ligase Modulator”. Clinical Pharmacology in Drug Development. 10 (5): 471–485. doi:10.1002/cpdd.869. PMC 8246954. PMID 32969202.
- Bjorklund, Chad C.; Kang, Jian; Amatangelo, Michael; Polonskaia, Ann; Katz, Mark; Chiu, Hsiling; et al. (April 2020). “Iberdomide (CC-220) is a potent cereblon E3 ligase modulator with antitumor and immunostimulatory activities in lenalidomide- and pomalidomide-resistant multiple myeloma cells with dysregulated CRBN”. Leukemia. 34 (4): 1197–1201. doi:10.1038/s41375-019-0620-8. ISSN 1476-5551. PMC 7214241. PMID 31719682.
- van de Donk, Niels W.C.J.; Popat, Rakesh; Larsen, Jeremy; Minnema, Monique C.; Jagannath, Sundar; Oriol, Albert; et al. (5 November 2020). “First Results of Iberdomide (IBER; CC-220) in Combination with Dexamethasone (DEX) and Daratumumab (DARA) or Bortezomib (BORT) in Patients with Relapsed/Refractory Multiple Myeloma (RRMM)”. Blood. 136 (Supplement 1): 16–17. doi:10.1182/blood-2020-137743. S2CID 228828103.
- Thieblemont, Catherine; Munoz, Javier; Tucci, Alessandra; Visco, Carlo; Cartron, Guillaume; Corradini, Paolo; et al. (15 November 2022). “Iberdomide (CC-220) Monotherapy or in Combination with an Anti-CD20 Monoclonal Antibody As Effective Therapy in Patients with Relapsed/Refractory Lymphoma: Early Results from a Phase 1/2 Study”. Blood. 140 (Supplement 1): 569–572. doi:10.1182/blood-2022-162559. S2CID 256795199.
- Lonial, Sagar; Amatangelo, Michael; Popat, Rakesh; Minnema, Monique C.; Zonder, Jeffrey A.; Larsen, Jeremy; et al. (13 November 2019). “Translational and Clinical Evidence of a Differentiated Profile for the Novel CELMoD, Iberdomide (CC-220)”. Blood. 134 (Supplement_1): 3119. doi:10.1182/blood-2019-124298. S2CID 209233746.
- Amatangelo, Michael; Bjorklund, Chad C.; Kang, Jian; Polonskaia, Ann; Viswanatha, Sridevi; Thakurta, Anjan (29 November 2018). “Iberdomide (CC-220) Has Synergistic Anti-Tumor and Immunostimulatory Activity Against Multiple Myeloma in Combination with Both Bortezomib and Dexamethasone, or in Combination with Daratumumab in Vitro”. Blood. 132 (Supplement 1): 1935. doi:10.1182/blood-2018-99-113383. S2CID 91382999.
- Lonial, Sagar; Popat, Rakesh; Hulin, Cyrille; Jagannath, Sundar; Oriol, Albert; Richardson, Paul G; et al. (November 2022). “Iberdomide plus dexamethasone in heavily pretreated late-line relapsed or refractory multiple myeloma (CC-220-MM-001): a multicentre, multicohort, open-label, phase 1/2 trial”. The Lancet Haematology. 9 (11): e822–e832. doi:10.1016/S2352-3026(22)00290-3. PMID 36209764. S2CID 252779185.
- Merrill, Joan T.; Werth, Victoria P.; Furie, Richard; van Vollenhoven, Ronald; Dörner, Thomas; Petronijevic, Milan; et al. (17 March 2022). “Phase 2 Trial of Iberdomide in Systemic Lupus Erythematosus”. New England Journal of Medicine. 386 (11): 1034–1045. doi:10.1056/NEJMoa2106535. PMID 35294813. S2CID 247499089.
- Feuerstein, Adam (14 August 2026). “FDA clears Bristol multiple myeloma therapy, marking debut of new drug class”. STAT. Retrieved 14 August 2026.
- World Health Organization (2018). “International nonproprietary names for pharmaceutical substances (INN): recommended INN: list 79”. WHO Drug Information. 32 (1). hdl:10665/330941.
- “U.S. FDA Grants Accelerated Approval to Bristol Myers Squibb’s First CELMoD Therapy Zenbexus, in Combination with Daratumumab and Hyaluronidase-fihj and Dexamethasone (ZDd) for Patients with Multiple Myeloma, as Early as First Relapse”. Bristol Myers Squibb (Press release). 13 August 2026. Retrieved 16 August 2026.
External links
- “Iberdomide ( Code – C129048 )”. EVS Explore.
- “Iberdomide Hydrochloride ( Code – C141516 )”. EVS Explore.
- Clinical trial number NCT04975997 for “Open-label Study Comparing Iberdomide, Daratumumab and Dexamethasone (IberDd) Versus Daratumumab, Bortezomib, and Dexamethasone (DVd) in Participants With Relapsed or Refractory Multiple Myeloma (RRMM) (EXCALIBER-RRMM)” at ClinicalTrials.gov
 | |
| Clinical data | |
|---|---|
| Trade names | Zenbexus |
| Other names | CC-220 |
| AHFS/Drugs.com | zenbexus |
| License data | US DailyMed: Iberdomide |
| Routes of administration | By mouth |
| Drug class | Cereblon-modulating protein degrader |
| ATC code | None |
| Legal status | |
| Legal status | US: ℞-only[1] |
| Identifiers | |
| IUPAC name | |
| CAS Number | 1323403-33-3as HCl: 1560678-63-8 |
| PubChem CID | 67335295as HCl: 72793904 |
| IUPHAR/BPS | 9618 |
| DrugBank | DB12101 |
| ChemSpider | 52085251 |
| UNII | 8V66F27X44as HCl: 79L3645KFI |
| KEGG | D11134as HCl: D11135 |
| ChEMBL | ChEMBL3989927 |
| Chemical and physical data | |
| Formula | C25H27N3O5 |
| Molar mass | 449.507 g·mol−1 |
| 3D model (JSmol) | Interactive imageas HCl: Interactive image |
| SMILES | |
| InChI | |
////////iberdomide, ANAX LABS, APPROVAL 2026, FDA 2026, Zenbexus, APPROVAL 2026, FDA 2026, Zenbexus, cc-220, cc 220, 8V66F27X44, 79L3645KFI
#iberdomide, #ANAX LABS, #APPROVAL 2026, #FDA 2026, #Zenbexus, #APPROVAL 2026, #FDA 2026, #Zenbexus, #cc-220, #cc 220, #8V66F27X44, #79L3645KFI
Gedatolisib
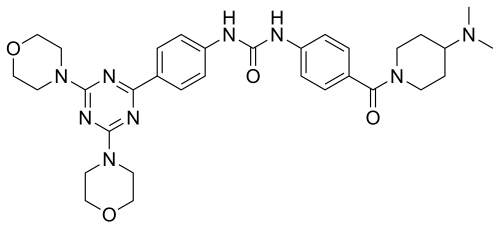
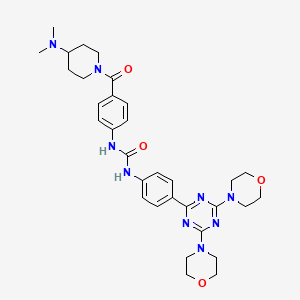
Gedatolisib
Approvals 3026, FDA 2026, 7/14/2026, Revtorpyk
| PF-05212384; PF-5212384; PKI-587 CAS 1197160-78-3 Chemical Formula: C32H41N9O4 Molecular Weight: 615.72 1-(4-{[4-(Dimethylamino)-1-piperidinyl]carbonyl}phenyl)-3-{4-[4,6-di(4-morpholinyl)-1,3,5-triazin-2-yl]phenyl}urea 3-{4-[bis(morpholin-4-yl)-1,3,5-triazin-2-yl]phenyl}-1-{4-[4-(dimethylamino)piperidine-1-carbonyl]phenyl}urea N-[4-[[4-(Dimethylamino)-1-piperidinyl]carbonyl]phenyl]-N’-[4-[4,6-di(4-morpholinyl)-1,3,5-triazin-2-yl]phenyl]urea гедатолисиб [Russian] [INN] غيداتوليسيب [Arabic] [INN] 吉达利塞 [Chinese] [INN] |
1-(4-(4-(Dimethylamino)piperidine-1-carbonyl)phenyl)-3-(4-(4,6-dimorpholino-1,3,5-triazin-2-yl)phenyl)urea
In combination with fulvestran, to treat hormone receptor-positive, human epidermal growth factor receptor 2-negative, locally advanced or metastatic breast cancer without a PIK3CA mutation detected following progression on or after treatment with at least one line of endocrine therapy in the metastatic setting
Gedatolisib, sold under the brand name Revtorpyk, is an anti-cancer drug used for the treatment of breast cancer.[1] It is under development by Celcuity, Inc. Gedatolisib is a kinase inhibitor.[1] The mechanism of action is accomplished by binding the different p110 catalytic subunit isoforms of PI3K and the kinase site of mTOR.[2] Gedatolisib is administered by intravenous infusion.[1]
Gedatolisib was approved for medical use in the United States in July 2026.[1][3]
Medical uses
Gedatolisib is indicated in combination with fulvestrant, with or without palbociclib, for the treatment of adults with hormone receptor (HR)-positive, human epidermal growth factor receptor 2 (HER2)-negative locally advanced or metastatic breast cancer without a PIK3CA mutation detected following progression on or after treatment with at least one line of endocrine therapy in the metastatic setting.[1]
- Synthesis of New Dialkyl 2,2′-[Carbonylbis(azanediyl)]dibenzoates via Curtius RearrangementDOI: 10.1055/s-0040-1706643Publication Date: 2021Publication Name: Synthesis
- A New One-Pot Three-Component Synthesis of 4-Aryl-6-cycloamino-1,3,5-triazin-2-amines under Microwave IrradiationDOI: 10.1055/a-1401-2795Publication Date: 2021Publication Name: Synthesis
PAT
https://patentscope.wipo.int/search/en/detail.jsf?docId=US42900900&_cid=P22-MRVGV9-51327-1
Example 76
Preparation of 1-(4-(4-(dimethylamino)piperidine-1-carbonyl)phenyl)-3-(4-(4,6-dimorpholino-1,3,5-triazin-2-yl)phenyl)urea
PATENT
https://patentscope.wipo.int/search/en/detail.jsf?docId=WO2010096619&_cid=P22-MRVGV9-51327-1
Scheme 1
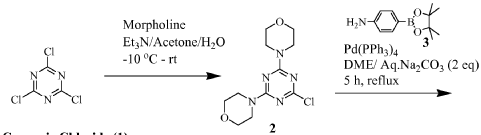
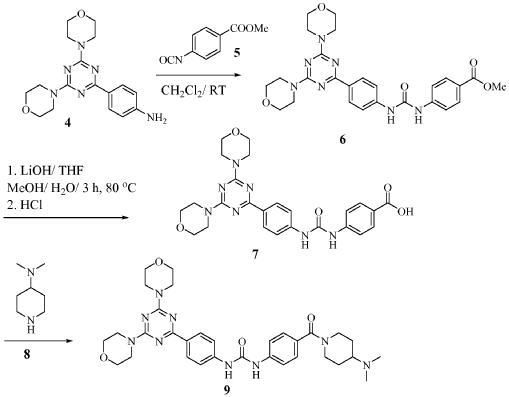
Preparation of 1-(4-(4-(dimethylamino) piperidine-1-carbonyl)phenyl-3-(4-(4,6- dimorpholino-1 ,3,5-triazine-2-yl)phenyl) urea (9)
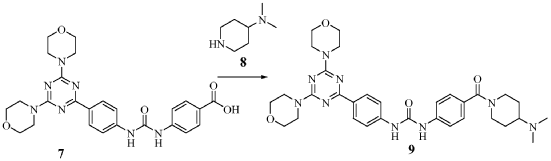
To a slurry of 4-(3-(4-(4,6-dimorpholino-1 ,3,5-triazine-2- yl)phenyl)ureido)benzoic acid (7, 45.5 g, 0.09 mol) in dry THF (1.6 L) heated to 50 0C was added N,N’-carbonyl diimidazole (28 g, 0.17 mol). The reaction mixture was heated for 2 hours and followed by dimethylaminopiperidine (8, 23.5 g, 0.18 mol) and stirred at 53 0C for 16 hours. The reaction mixture was cooled to the room temperature and filtered. The cake was washed with 2-propanol and air-dried to give 97 % pure white powder in 88% yield (49.2 g, 0.08 mol). To the solids stirred in dimethyl acetamide (DMAC, 165 ml) at 70° C for 1 hour was added 2-propanol (640 ml) and the mixture was stirred at 65 0C for additional 1 hour. The solids were filtered, washed with 2-propanol and dried in a vacuum oven at 700C for 16 hour to give crystalline white powder (45 g) with >99% purity. The above-mentioned work up process and crystallization procedure gave a Pd residue of 20 ppm. Alternate procedures for the formation of 1-(4-(4-(dimethylamino) piperidine-1 – carbonyl)phenyl-3-(4-(4,6-dimorpholino-1 ,3,5-triazine-2-yl)phenyl) urea (9)
To the solution of 4-(4,6-dimorpholin-4-yl-1 ,3,5-triazin-2-yl) aniline (4, 18 g, 0.052 mol) in dichloromethane (300 ml) was added methyl 4-isocyanato benzoate (5, 10.5 g, 0.061 mol) and the reaction mixture was stirred for 5 hours. The separated solids were filtered, washed with ether and air dried to give beige solids (21 g, 0.04 mol). Yield 77%. 90 % pure by HPLC; Mass: 520.1 (M+H). Preparation of 4-(3-(4-(4,6-dimorpholino-1 ,3,5-triazine-2-yl)phenyl)ureido) benzoic acid (7)
The mixture of methyl 4-(3-(4-(4,6-dimorpholino-1 ,3,5-triazine-2-yl)ureido)benzoate (6, 21 g, 0.04mol) and lithium hydroxide monohydrate (3.8 g, 0.09 mol) in THF (120 ml), MeOH (60 ml), and water (60 ml) was heated at 80 0C for 3 hours. The dark brown solution was cooled to room temperature and made acidic with concentrated HCI. The solids were filtered, washed with water, washed with acetone , washed with ether, and dried in a vacuum oven at 60 0C for 48 hours to give off white solids of 4-(3-(4-(4,6-dimorpholino-1 ,3,5-triazine-2-yl)phenyl)ureido) benzoic acid (19.2 g, 0.038 mol). Mass: 506.3 (M+H)+; Yield.94%. 1 -(4-(4-(dimethylamino) piperidine-1 -carbonyl)phenyl-3-(4-(4,6-dimorpholino- 1 ,3,5-triazine-2-yl)phenyl) urea (9)
The suspension of 4-(3-(4-(4,6-dimorpholino-1 ,3,5-triazine-2-yl)phenyl)ureido) benzoic acid (7, 17 g, 33.66 mmol) and N-(3-dimethylaminopropyl)ethyl carbodiimide hydrochloride (9.5 g, 49.5 mmol) in THF (200 ml) and acetonitrile (50 ml) was stirred for 10 min and followed by addition of 1-hydroxybenzotriazole hydrate (6.4 g, 47.88 mmol). The reaction mixture was stirred for 30 min and 4-dimethylaminopiperidine (8, 8.86 g, 69.2 mmol) was added by drops. After being stirred for additional 16 hours, the reaction mixture was concentrated to min. The solids were filtered and washed thoroughly with water (very fine suspension). The cake was slurred in hot ethanol, filtered and dried in a vacuum oven at 68 0C for 16 hours to give off white solids (10.3 g, 16.77 mmol). M. p. 238-240 0C. 99 % pure. Mass: 616.3 (M+H)+; Yield 50 %.
PATENT
WO 2009143317
WO 2010096619
WO 2012148540
WO 2014151147
PATENT
US 20170119778
PAPER
Journal of Medicinal Chemistry (2010), 53(6), 2636-2645
http://pubs.acs.org/doi/abs/10.1021/jm901830p
J. Med. Chem., 2010, 53 (6), pp 2636–2645
DOI: 10.1021/jm901830p
Abstract
The PI3K/Akt signaling pathway is a key pathway in cell proliferation, growth, survival, protein synthesis, and glucose metabolism. It has been recognized recently that inhibiting this pathway might provide a viable therapy for cancer. A series of bis(morpholino-1,3,5-triazine) derivatives were prepared and optimized to provide the highly efficacious PI3K/mTOR inhibitor 1-(4-{[4-(dimethylamino)piperidin-1-yl]carbonyl}phenyl)-3-[4-(4,6-dimorpholin-4-yl-1,3,5-triazin-2-yl)phenyl]urea 26 (PKI-587). Compound 26 has shown excellent activity in vitro and in vivo, with antitumor efficacy in both subcutaneous and orthotopic xenograft tumor models when administered intravenously. The structure−activity relationships and the in vitro and in vivo activity of analogues in this series are described.
Preparation of 1-(4-{[4-(Dimethylamino)piperidin-1-yl]carbonyl}phenyl)-3-[4-(4,6-dimorpholin-4- yl-1,3,5-triazin-2-yl)phenyl]urea (26)
MS (ESI) m/z = 616.7. HRMS: calcd for C32H41N9O4 + H+, 616.335 43; found (ESI-FTMS, [M + H]+), 616.334 24. Purity by analytical HPLC 99.3%. (Prodigy ODS3, 0.46 cm × 15 cm, 20 min gradient acetonitrile in water, trifluoroacetic acid, detector wavelengths, 215 and 254 nm.) 1H NMR (DMSO-d6) δ 1.29−1.36 (m, 6H), 2.6 (m, 4H), 2.9 (m,1H), 3.3 (m, 4H), 3.6 (m, 8H), 3.7 (m, 8H), 7.3 (d, J = 8.3 Hz, 2H), 7.51−7.57 (m, 4H), 8.3 (d, J = 8.3 Hz 2H), 8.9 (s, 1H), 9.0 (s, 1H) ppm. Anal. Calcd for C32H41N9O4: C 62.42%, H 6.71%, N 20.47%. Found: C 62.34%, H 6.67%, N 20.39%.
PAPER
Bioorganic & Medicinal Chemistry Letters (2011), 21(16), 4773-4778.
http://www.sciencedirect.com/science/article/pii/S0960894X11008468
PAPER
New and Practical Synthesis of Gedatolisib
http://pubs.acs.org/doi/10.1021/acs.oprd.7b00298
Org. Process Res. Dev., Article ASAP
DOI: 10.1021/acs.oprd.7b00298
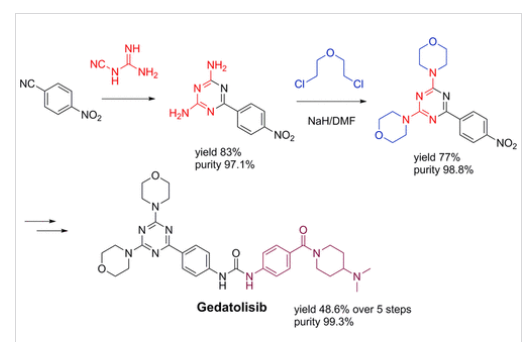
Abstract
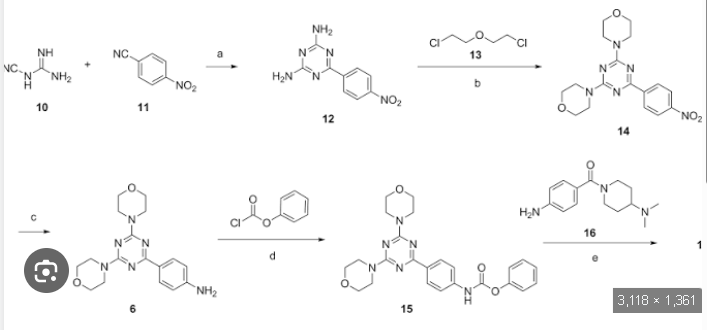
A new, practical, and convergent synthetic route of gedatolisib, an antitumor agent, is developed on a hectogram scale which avoids the Pd coupling method. The key step is adopting 6-(4-nitrophenyl)-1,3,5-triazine-2,4-diamine and 2,2′-dichlorodiethyl ether to prepare the key 4,4′-(6-(4-nitrophenyl)-1,3,5-triazine-2,4-diyl)dimorpholine in 77% yield and 98.8% purity. Gedatolisib is obtained in 48.6% yield over five simple steps and 99.3% purity (HPLC). Purification methods of the intermediates and the final product involved in the route are given.
off-white solid. 1H NMR (400 MHz, DMSO-d6): δ 1.46 (brs, 2H), 1.89 (brs, 2H), 2.29 (s, 6H), 2.94 (brs, 2H), 3.76 (m, 8H), 3.89 (m, 8H), 7.09 (d, J = 8.4 Hz, 2H), 7.20 (d, J = 8.4 Hz, 2H), 7.50 (d, J = 8.7 Hz, 2H), 8.28 (s, 1H), 8.31 (d, J = 8.6 Hz, 2H), 8.48 (s, 1H). ESI-MS (m/z) 615.9 (M + H). HPLC conditions: Column: Agilent Eclipse XDB-C18 (250 mm × 4.6 mm × 5 μm); Detection: 254 nm; Flow rate: 0.8 mL/min; Temperature: 30 °C; Injection load: 1 μL; Solvent: MeOH; Concentration: 0.5 mg/mL; Run time: 20 min; Mobile phase A: water; Mobile phase B: MeOH/TEA = 100:0.1; Gradient program: time (min): 20; % of mobile phase A: 10; % of mobile phase B: 90; tR = 2.598 min, purity: 99.34%
- Zhao, X.; Tan, Q.; Zhang, Z.; Zhao, Y. Med. Chem. Res. 2014, 23, 5188– 5196 DOI: 10.1007/s00044-014-1084-z
- Khafizova, G.; Potoski, J. R. PCT Int. Appl. WO 2010096619, 2010.
- Venkatesan, A. M.; Chen, Z.; Dehnhardt, C. M.; Dos Santos, O.; Delos Santos, E. G.; Zask, A.; Verheijen, J. C.; Kaplan, J. A.; Richard, D. J.; Ayral-Kaloustian, S.; Mansour, T. S.; Gopalsamy, A.; Curran, K. J.; Shi, M. PCT Int. Appl. WO 2009143317, 2009.
REFERENCES
1: Gedaly R, Galuppo R, Musgrave Y, Angulo P, Hundley J, Shah M, Daily MF, Chen C, Cohen DA, Spear BT, Evers BM. PKI-587 and sorafenib alone and in combination on inhibition of liver cancer stem cell proliferation. J Surg Res. 2013 Nov;185(1):225-30. doi: 10.1016/j.jss.2013.05.016. Epub 2013 May 25. PubMed PMID: 23769634.
2: Gedaly R, Angulo P, Hundley J, Daily MF, Chen C, Evers BM. PKI-587 and sorafenib targeting PI3K/AKT/mTOR and Ras/Raf/MAPK pathways synergistically inhibit HCC cell proliferation. J Surg Res. 2012 Aug;176(2):542-8. doi: 10.1016/j.jss.2011.10.045. Epub 2011 Nov 21. PubMed PMID: 22261591.
3: Dehnhardt CM, Venkatesan AM, Chen Z, Delos-Santos E, Ayral-Kaloustian S, Brooijmans N, Yu K, Hollander I, Feldberg L, Lucas J, Mallon R. Identification of 2-oxatriazines as highly potent pan-PI3K/mTOR dual inhibitors. Bioorg Med Chem Lett. 2011 Aug 15;21(16):4773-8. doi: 10.1016/j.bmcl.2011.06.063. Epub 2011 Jun 21. PubMed PMID: 21763134.
4: Mallon R, Feldberg LR, Lucas J, Chaudhary I, Dehnhardt C, Santos ED, Chen Z, dos Santos O, Ayral-Kaloustian S, Venkatesan A, Hollander I. Antitumor efficacy of PKI-587, a highly potent dual PI3K/mTOR kinase inhibitor. Clin Cancer Res. 2011 May 15;17(10):3193-203. doi: 10.1158/1078-0432.CCR-10-1694. Epub 2011 Feb 15. PubMed PMID: 21325073.
5: Venkatesan AM, Chen Z, dos Santos O, Dehnhardt C, Santos ED, Ayral-Kaloustian S, Mallon R, Hollander I, Feldberg L, Lucas J, Yu K, Chaudhary I, Mansour TS. PKI-179: an orally efficacious dual phosphatidylinositol-3-kinase (PI3K)/mammalian target of rapamycin (mTOR) inhibitor. Bioorg Med Chem Lett. 2010 Oct 1;20(19):5869-73. doi: 10.1016/j.bmcl.2010.07.104. Epub 2010 Jul 30. PubMed PMID: 20797855.
6: Venkatesan AM, Dehnhardt CM, Delos Santos E, Chen Z, Dos Santos O, Ayral-Kaloustian S, Khafizova G, Brooijmans N, Mallon R, Hollander I, Feldberg L, Lucas J, Yu K, Gibbons J, Abraham RT, Chaudhary I, Mansour TS. Bis(morpholino-1,3,5-triazine) derivatives: potent adenosine 5′-triphosphate competitive phosphatidylinositol-3-kinase/mammalian target of rapamycin inhibitors: discovery of compound 26 (PKI-587), a highly efficacious dual inhibitor. J Med Chem. 2010 Mar 25;53(6):2636-45. doi: 10.1021/jm901830p. PubMed PMID: 20166697.
ADVERTISEMENT
ANAX LABORATORIES
WEBSITE https://www.anaxlab.com/
Discovery Solutions, Supporting the chemistry needs of clients in the Medical, Analytical and Bio Sciences
Development Solutions, Developing from Lab scale to PR&D, Kilo Scale-ups and Commercial Scales
SEE MORE………Integrated Solutions, Manufacturing Solutions, Products,
Can’t Find? Let’s Connect
Phone : +91 897704 2010 / +91 9177075735, Email : info@anaxlab.com
#MedicinalChemistry, #DrugDiscovery, #OrganicSynthesis, #ChemicalLibrary, #BuildingBlocks, #SARStudies, #ChemistryInnovation, #medchem, #Drugdevelopment, #Biotech, #Biotechnology, #AnaxLaboratories, #Pharma
AS ON FEB2026 4.574 LAKHS VIEWS ON BLOG WORLDREACH AVAILABLEFOR YOUR ADVERTISEMENT
join me on Linkedin
Anthony Melvin Crasto Ph.D – India | LinkedIn
join me on Researchgate
RESEARCHGATE
join me on Facebook
Anthony Melvin Crasto Dr. | Facebook
join me on twitter
Anthony Melvin Crasto Dr. | twitter
+919321316780 call whatsaapp
EMAIL. amcrasto@gmail.com
References
- PRMT5 inhibitors and uses thereofPublication Number: US-12448388-B2Grant Date: 2025-10-21
- KRAS G12D modulating compoundsPublication Number: US-12448400-B2Grant Date: 2025-10-21
- TRIAZINE COMPOUNDS AS P13 KINASE AND MTOR INHIBITORSPublication Number: PT-2294072-TPriority Date: 2008-05-23
- TRIASINE UNITS AS P13 KINASE INHIBITORS AND MOTORPublication Number: ME-01111-BPriority Date: 2008-05-23
- HER2 mutation inhibitorsPublication Number: US-12447153-B2Grant Date: 2025-10-21
- Anti-hiv compoundsPublication Number: US-2025326779-A1
- Aryl aminopyrimidines as dual MerTK and TYRO3 inhibitors and methods thereofPublication Number: US-12448365-B2Grant Date: 2025-10-21
References
- https://celcuity.com/revtorpyk/REVTORPYK_PI_2026.pdf
- Dehnhardt CM, Venkatesan AM, Chen Z, Delos-Santos E, Ayral-Kaloustian S, Brooijmans N, et al. (August 2011). “Identification of 2-oxatriazines as highly potent pan-PI3K/mTOR dual inhibitors”. Bioorganic & Medicinal Chemistry Letters. 21 (16): 4773–8. doi:10.1016/j.bmcl.2011.06.063. PMID 21763134.
- “FDA approves gedatolisib with fulvestrant, with or without palbociclib, for HR-positive, HER2-negative locally advanced or metastatic breast cancer”. U.S. Food and Drug Administration (FDA). 14 July 2026. Retrieved 20 July 2026.
 This article incorporates text from this source, which is in the public domain.
- Sabatini DM (November 2017). “Twenty-five years of mTOR: Uncovering the link from nutrients to growth”. Proceedings of the National Academy of Sciences of the United States of America. 114 (45): 11818–11825. Bibcode:2017PNAS..11411818S. doi:10.1073/pnas.1716173114. PMC 5692607. PMID 29078414.
- Tian T, Li X, Zhang J (February 2019). “mTOR Signaling in Cancer and mTOR Inhibitors in Solid Tumor Targeting Therapy”. International Journal of Molecular Sciences. 20 (3): 755. doi:10.3390/ijms20030755. PMC 6387042. PMID 30754640.
- Hua H, Kong Q, Zhang H, Wang J, Luo T, Jiang Y (July 2019). “Targeting mTOR for cancer therapy”. Journal of Hematology & Oncology. 12 (1) 71. doi:10.1186/s13045-019-0754-1. PMC 6612215. PMID 31277692.
- Vanhaesebroeck B, Perry MW, Brown JR, André F, Okkenhaug K (October 2021). “PI3K inhibitors are finally coming of age”. Nature Reviews. Drug Discovery. 20 (10): 741–769. doi:10.1038/s41573-021-00209-1. PMC 9297732. PMID 34127844. S2CID 235437841.
- Millis SZ, Ikeda S, Reddy S, Gatalica Z, Kurzrock R (December 2016). “Landscape of Phosphatidylinositol-3-Kinase Pathway Alterations Across 19 784 Diverse Solid Tumors”. JAMA Oncology. 2 (12): 1565–1573. doi:10.1001/jamaoncol.2016.0891. PMID 27388585.
- Anderson EJ, Mollon LE, Dean JL, Warholak TL, Aizer A, Platt EA, et al. (2020). “A Systematic Review of the Prevalence and Diagnostic Workup of PIK3CA Mutations in HR+/HER2- Metastatic Breast Cancer”. International Journal of Breast Cancer. 2020 3759179. doi:10.1155/2020/3759179. PMC 7322582. PMID 32637176.
- Clinical trial number NCT01420081 for “A Study Of Two Dual PI3K/mTOR Inhibitors, PF-04691502 And PF-05212384 In Patients With Recurrent Endometrial Cancer” at ClinicalTrials.gov
- Clinical trial number NCT01925274 for “A Study Of PF-05212384 Plus Irinotecan Vs Cetuximab Plus Irinotecan In Patients With KRAS And NRAS Wild Type Metastatic Colorectal Cancer” at ClinicalTrials.gov
- Clinical trial number NCT02438761 for “PF-05212384 (PKI-587) for t-AML/MDS or de Novo Relapsed or Refractory Acute Myeloid Leukemia (AML)” at ClinicalTrials.gov
- Clinical trial number NCT03698383 for “Phase II Study of Herzuma® Plus Gedatolisib in Patients With HER-2 Positive Metastatic Breast Cancer” at ClinicalTrials.gov
- Clinical trial number NCT03911973 for “Gedatolisib Plus Talazoparib in Advanced Triple Negative or BRCA1/2 Positive, HER2 Negative Breast Cancers” at ClinicalTrials.gov
- Clinical trial number NCT03065062 for “Study of the CDK4/6 Inhibitor Palbociclib (PD-0332991) in Combination With the PI3K/mTOR Inhibitor Gedatolisib (PF-05212384) for Patients With Advanced Squamous Cell Lung, Pancreatic, Head & Neck and Other Solid Tumors” at ClinicalTrials.gov
- Clinical trial number NCT02626507 for “Phase I Study of Combination of Gedatolisib With Palbociclib and Faslodex in Patients With ER+/HER2- Breast Cancer” at ClinicalTrials.gov
- “Celcuity Announces FDA Approval of Revtorpyk (gedatolisib) for the Treatment of HR+/HER2-, PIK3CA Wild-Type Locally Advanced or Metastatic Breast Cancer” (Press release). Celcuity. 14 July 2026. Retrieved 20 July 2026 – via GlobeNewswire.
- World Health Organization (2015). “International nonproprietary names for pharmaceutical substances (INN): recommended INN: list 73”. WHO Drug Information. 29 (1). hdl:10665/331088.
External links
- Clinical trial number NCT05501886 for “Gedatolisib Plus Fulvestrant With or Without Palbociclib vs Standard-of-Care for the Treatment of Patients With Advanced or Metastatic HR+/HER2- Breast Cancer (VIKTORIA-1) (VIKTORIA-1)” at ClinicalTrials.gov
 | |
| Clinical data | |
|---|---|
| Trade names | Revtorpyk |
| Other names | PF-05212384; PKI-587 |
| AHFS/Drugs.com | revtorpyk |
| License data | US DailyMed: Gedatolisib |
| Routes of administration | Intravenous infusion |
| Drug class | Antineoplastic |
| ATC code | None |
| Legal status | |
| Legal status | US: ℞-only[1] |
| Identifiers | |
| IUPAC name | |
| CAS Number | 1197160-78-3 |
| PubChem CID | 44516953 |
| IUPHAR/BPS | 7940 |
| DrugBank | DB11896 |
| ChemSpider | 24644946 |
| UNII | 96265TNH2R |
| KEGG | D10635 |
| ChEMBL | ChEMBL592445 |
| CompTox Dashboard (EPA) | DTXSID40152557  |
| Chemical and physical data | |
| Formula | C32H41N9O4 |
| Molar mass | 615.739 g·mol−1 |
| 3D model (JSmol) | Interactive image |
| SMILES | |
/////////Gedatolisib, anax labs, approvals 3026, FDA 2026, PF 05212384, PF 5212384, PKI-587, PF-05212384, PF-5212384, PKI 587, gedatolisib, antitumor agent, PHASE 3, PFIZER, гедатолисиб , غيداتوليسيب , 吉达利塞 , 96265TNH2R
O=C(NC1=CC=C(C2=NC(N3CCOCC3)=NC(N4CCOCC4)=N2)C=C1)NC5=CC=C(C(N6CCC(N(C)C)CC6)=O)C=C5
Journal of Medicinal Chemistry (2017), 60(17), 7524-7538 PQR 309
#Gedatolisib, #anax labs, #approvals 3026, #FDA 2026, #PF 05212384, #PF 5212384, #PKI-587, #PF-052123842, #PF-5212384; #PKI 587, #gedatolisib, #antitumor agent, #PHASE 3, #PFIZER, #гедатолисиб , #غيداتوليسيب , #吉达利塞 , #96265TNH2R
Foselutoclax
Foselutoclax
CAS 2271269-01-1
MF C53H59ClF3N6O10PS3 MW 1159.7 g/mol
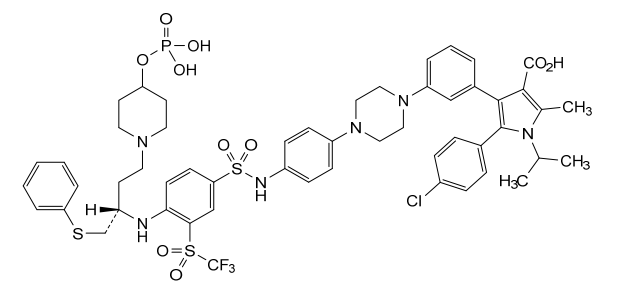
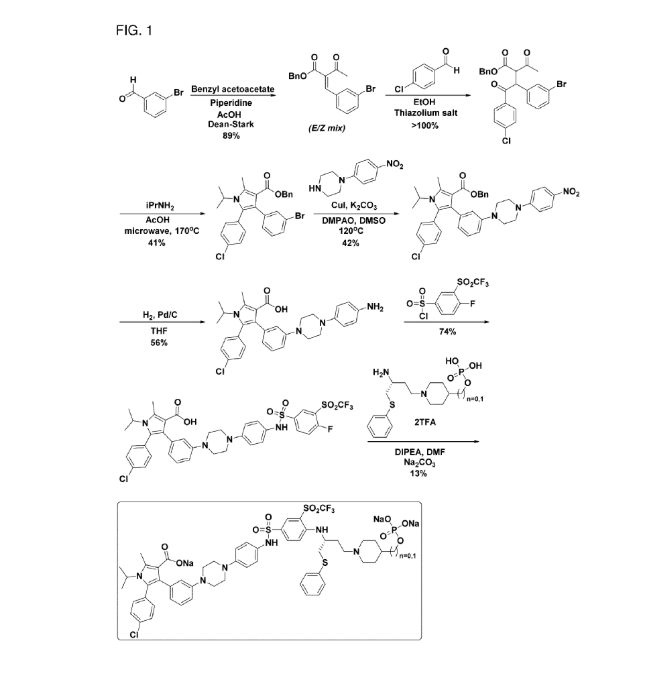
(10R)-14-chloro-25-methyl-7,7-dioxo-10-[(phenylsulfanyl)methyl]-134-(phosphonooxy)-21-(propan-2-yl)-83-(trifluoromethanesulfonyl)-21H-7λ6-thia-6,9-diaza-4(1,4)-piperazina-13(1)-piperidina-2(2,3)-pyrrola-1(1),3(1,3),5,8(1,4)-tetrabenzenatridecaphane-24-carboxylic acid
5-(4-chlorophenyl)-2-methyl-4-[3-[4-[4-[[4-[[(2R)-1-phenylsulfanyl-4-(4-phosphonooxypiperidin-1-yl)butan-2-yl]amino]-3-(trifluoromethylsulfonyl)phenyl]sulfonylamino]phenyl]piperazin-1-yl]phenyl]-1-propan-2-ylpyrrole-3-carboxylic acid
B-cell lymphoma 2 (Bcl-2) inhibitor, antineoplastic, VT53CL5GES, UBX 1325
Foselutoclax is an investigational new drug that is being evaluated for the treatment of age-related eye diseases, particularly diabetic macular edema (DME) and wet age-related macular degeneration (AMD). Developed by Unity Biotechnology, this senolytic compound acts as a potent inhibitor of Bcl-xL, a protein that senescent cells rely on for survival.[1] Foselutoclax is designed to selectively eliminate senescent cells in the retina, potentially addressing the underlying causes of vision loss in these conditions.[2]
- Assess the Efficacy and Safety of Repeat Intravitreal Injections of Foselutoclax (UBX1325) in Patients With DME (ASPIRE)CTID: NCT06011798Phase: Phase 2Status: CompletedDate: 2025-08-05
- Safety, Tolerability and Evidence of Activity Study of UBX1325 in Patients With Diabetic Macular Edema (BEHOLD)CTID: NCT04857996Phase: Phase 2Status: CompletedDate: 2024-05-16
- Safety and Tolerability Study of UBX1325 in Patients With Diabetic Macular Edema or Neovascular Age-Related Macular DegenerationCTID: NCT04537884Phase: Phase 1Status: CompletedDate: 2022-03-10
REF
- Therapeutic targeting of cellular senescence in diabetic macular edema: preclinical and phase 1 trial resultsPublication Name: Nature MedicinePublication Date: 2024-02PMID: 38321220DOI: 10.1038/s41591-024-02802-4
- Senolytics in the treatment of diabetic retinopathyPublication Name: Frontiers in PharmacologyPublication Date: 2022-08-26PMCID: PMC9462063PMID: 36091769DOI: 10.3389/fphar.2022.896907
- Senolytic drugs: from discovery to translationPublication Name: Journal of Internal MedicinePublication Date: 2020-08-04PMCID: PMC7405395PMID: 32686219DOI: 10.1111/joim.13141
- bcl-x, a bcl-2-related gene that functions as a dominant regulator of apoptotic cell deathPublication Name: CellPublication Date: 1993-08-27PMID: 8358789DOI: 10.1016/0092-8674(93)90508-n
PAT
Treatment of Lung Diseases Using Pharmaceutical Agents that Eliminate Senescent Cells
Publication Number: US-2020354336-A9
Priority Date: 2017-08-11
- Senescent Cells and for Treating CancerPublication Number: US-2022017485-A1Priority Date: 2018-06-13
- Acyl sulfonamides that are bcl family antagonists for use in clinical management of conditions caused or mediated by senescent cells and for treating cancerPublication Number: EP-4335516-A2Priority Date: 2018-06-13
- Methods of Inhibiting Pathological AngiogenesisPublication Number: US-2020253991-A1Priority Date: 2017-10-31
- Methods of inhibiting pathological angiogenesisPublication Number: US-11129838-B2Priority Date: 2017-10-31Grant Date: 2021-09-28
- Treatment of Lung Diseases Using Pharmaceutical Agents that Eliminate Senescent CellsPublication Number: US-2020199103-A1Priority Date: 2017-08-11
SYN
https://patentscope.wipo.int/search/en/detail.jsf?docId=US279621490&_cid=P21-MGPXU3-15237-1
SYN
https://patentscope.wipo.int/search/en/detail.jsf?docId=US421382898&_cid=P21-MGPXWE-19244-1
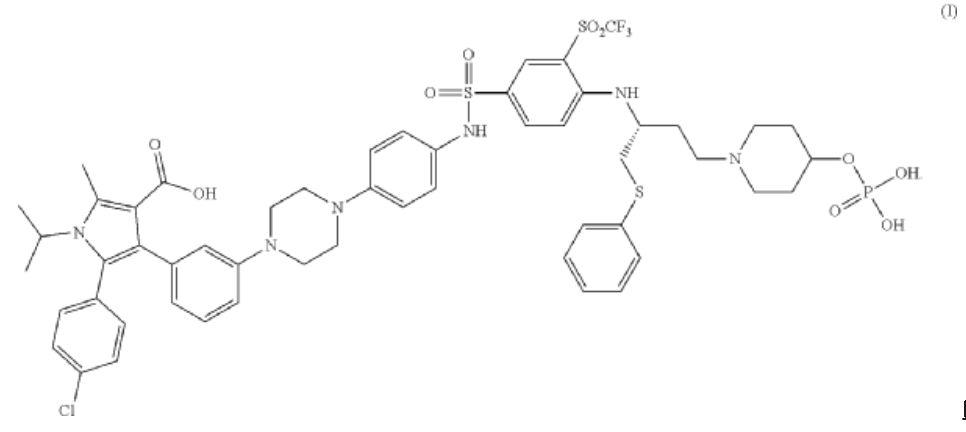
A crystalline solid meglumine salt of of (R)-5-(4-chlorophenyl)-1-isopropyl-2-methyl-4-(3-(4-(4-((4-((1-(phenylthio)-4-(4-((phosphonooxy)methyl)piperidin-1-yl)butan-2-yl)amino)-3-((trifluoromethyl)sulfonyl)phenyl)sulfonamido)phenyl)piperazin-1-yl)phenyl)-1H-pyrrole-3-carboxylic acid, the compound of Formula I:
SYN
https://patentscope.wipo.int/search/en/detail.jsf?docId=US348024244&_cid=P21-MGPXWE-19244-1

AS ON JUNE2025 4.45 LAKHS VIEWS ON BLOG WORLDREACH AVAILABLEFOR YOUR ADVERTISEMENT
join me on Linkedin
Anthony Melvin Crasto Ph.D – India | LinkedIn
join me on Researchgate
RESEARCHGATE
join me on Facebook
Anthony Melvin Crasto Dr. | Facebook
join me on twitter
Anthony Melvin Crasto Dr. | twitter
+919321316780 call whatsaapp
EMAIL. amcrasto@gmail.com
……
| Clinical data | |
|---|---|
| Other names | UBX1325 |
| Identifiers | |
| IUPAC name | |
| CAS Number | 2271269-01-1 |
| PubChem CID | 147562879 |
| IUPHAR/BPS | 13366 |
| ChemSpider | 115277082 |
| UNII | VT53CL5GES |
| Chemical and physical data | |
| Formula | C53H59ClF3N6O10PS3 |
| Molar mass | 1159.69 g·mol−1 |
| 3D model (JSmol) | Interactive image |
| SMILES | |
| InChI | |
References
- Crago SM (22 June 2023). “Design for Phase 2B ASPIRE Study of UBX1325 for DME announced by UNITY”. Modern Retina. Archived from the original on 13 August 2024.
- Macha N, Yu M, Sapieha P, Klier S, Ghosh A, White L, et al. (September 2024). “Multifocal Electroretinography Changes after UBX1325 (Foselutoclax) Treatment in Neovascular Age-Related Macular Degeneration”. Journal of Clinical Medicine. 13 (18): 5540. doi:10.3390/jcm13185540. PMC 11433175. PMID 39337030.
//////////foselutoclax, antineoplastic, VT53CL5GES, UBX 1325
Ezobresib
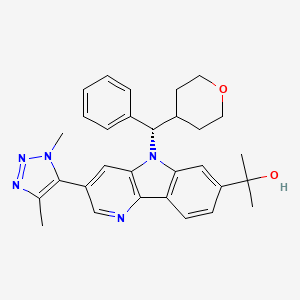
Ezobresib
CAS 1800340-40-2
MF C30H33N5O2 MW 495.6 g/mol
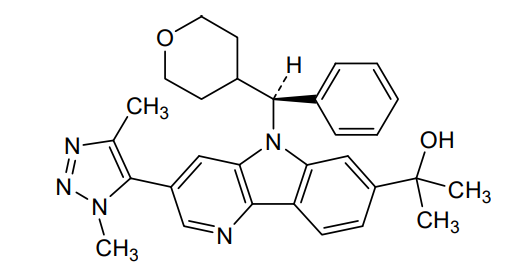
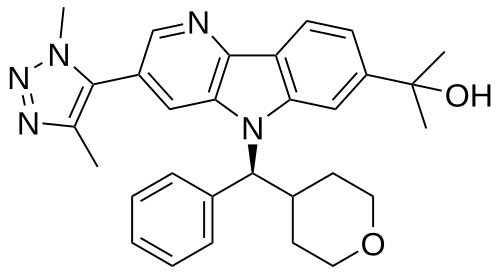
2-{3-(1,4-dimethyl-1H-1,2,3-triazol-5-yl)-5-[(S)-(oxan-4-yl)(phenyl)methyl]-5H-pyrido[3,2-b]indol-7-yl}propan-2-ol
bromodomain and extra-terminal motif (BET) inhibitor,
antineoplastic, BMS-986158, BMS 986158, Bristol Myers Squibb, antineoplastic, UNII-X8BW0MQ5PI
2-[3-(3,5-dimethyltriazol-4-yl)-5-[(S)-oxan-4-yl(phenyl)methyl]pyrido[3,2-b]indol-7-yl]propan-2-ol
Ezobresib is an investigational new drug that has been evaluated for the treatment of cancer. It inhibits Bromodomain and Extra-Terminal domain (BET) proteins, with potential antineoplastic activity.[1] Developed by Bristol Myers Squibb, this therapeutic agent has been studied for its efficacy in treating various cancers, including solid tumors and hematological malignancies.[2] Despite showing promise in early-phase clinical trials, recent developments suggest that Bristol Myers Squibb has decided to discontinue further development of ezobresib.[3]
BMS-986158 is under investigation in clinical trial NCT02419417 (Study of BMS-986158 in Subjects With Select Advanced Cancers).
Ezobresib is an inhibitor of the Bromodomain (BRD) and Extra-Terminal domain (BET) family of proteins, with potential antineoplastic activity. Upon administration, ezobresib binds to the acetyl-lysine binding site in the BRD of BET proteins, thereby preventing the interaction between BET proteins and acetylated histones. This disrupts chromatin remodeling and prevents the expression of certain growth-promoting genes, resulting in an inhibition of tumor cell growth. BET proteins (BRD2, BRD3, BRD4 and BRDT) are transcriptional regulators that bind to acetylated lysines on the tails of histones H3 and H4, and regulate chromatin structure and function; they play an important role in the modulation of gene expression during development and cellular growth
SYN
https://patentscope.wipo.int/search/en/detail.jsf?docId=US206490064&_cid=P21-MGLNPO-16484-1
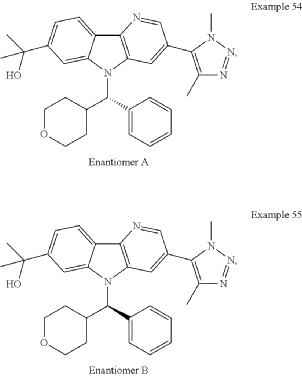
Examples 54 & 55
2-[3-(Dimethyl-1H-1,2,3-triazol-5-yl)-5-[oxan-4-yl(phenyl)methyl]-5H-pyrido[3,2-b]indol-7-yl]propan-2-ol
Step 1: 2-Chloro-5-(1,4-dimethyl-1H-1,2,3-triazol-5-yl)pyridin-3-amine
Step 2: Methyl 3-((2-chloro-5-(1,4-dimethyl-1H-1,2,3-triazol-5-yl)pyridin-3-yl)amino)benzoate
Step 3: Methyl 3-(1,4-dimethyl-1H-1,2,3-triazol-5-yl)-5H-pyrido[3,2-b]indole-7-carboxylate
Alternate synthesis of Methyl 3-(1,4-dimethyl-1H-1,2,3-triazol-5-yl)-5H-pyrido[3,2-b]indole-7-carboxylate
Step 4: Methyl 3-(1,4-dimethyl-1H-1,2,3-triazol-5-yl)-5-(phenyl(tetrahydro-2H-pyran-4-yl)methyl)-5H-pyrido[3,2-b]indole-7-carboxylate
Step 5: 2-[3-(Dimethyl-1H-1,2,3-triazol-5-yl)-5-[oxan-4-yl(phenyl)methyl]-5H-pyrido[3,2-b]indol-7-yl]propan-2-ol
Alternate Synthesis of Examples 54
2-[3-(Dimethyl-1H-1,2,3-triazol-5-yl)-5-[oxan-4-yl(phenyl)methyl]-5H-pyrido[3,2-b]indol-7-yl]propan-2-ol
Step 1: 2-Chloro-5-(1,4-dimethyl-1H-1,2,3-triazol-5-yl)pyridin-3-amine
Step 2: Methyl 3-((2-chloro-5-(1,4-dimethyl-1H-1,2,3-triazol-5-yl)pyridin-3-yl)amino)benzoate
Step 3: Methyl 3-(1,4-dimethyl-1H-1,2,3-triazol-5-yl)-5H-pyrido[3,2-b]indole-7-carboxylate
Alternate synthesis of Methyl 3-(1,4-dimethyl-1H-1,2,3-triazol-5-yl)-5H-pyrido[3,2-b]indole-7-carboxylate
Step 4: Methyl 3-(1,4-dimethyl-1H-1,2,3-triazol-5-yl)-5-(phenyl(tetrahydro-2H-pyran-4-yl)methyl)-5H-pyrido[3,2-b]indole-7-carboxylate
Step 5: 2-[3-(Dimethyl-1H-1,2,3-triazol-5-yl)-5-[oxan-4-yl(phenyl)methyl]-5H-pyrido[3,2-b]indol-7-yl]propan-2-ol
Alternate Synthesis of Examples 54
2-[3-(Dimethyl-1H-1,2,3-triazol-5-yl)-5-[oxan-4-yl(phenyl)methyl]-5H-pyrido[3,2-b]indol-7-yl]propan-2-ol
Step 1: (S)-methyl 3-(1,4-dimethyl-1H-1,2,3-triazol-5-yl)-5-(phenyl(tetrahydro-2H-pyran-4-yl)methyl)-5H-pyrido[3,2-b]indole-7-carboxylate
Step 2. (S)-2-[3-(Dimethyl-1H-1,2,3-triazol-5-yl)-5-[oxan-4-yl(phenyl)methyl]-5H-pyrido[3,2-b]indol-7-yl]propan-2-ol
Step 1: (S)-methyl 3-(1,4-dimethyl-1H-1,2,3-triazol-5-yl)-5-(phenyl(tetrahydro-2H-pyran-4-yl)methyl)-5H-pyrido[3,2-b]indole-7-carboxylate
Step 2. (S)-2-[3-(Dimethyl-1H-1,2,3-triazol-5-yl)-5-[oxan-4-yl(phenyl)methyl]-5H-pyrido[3,2-b]indol-7-yl]propan-2-ol
PATENT
LIT
- BLM overexpression as a predictive biomarker for CHK1 inhibitor response in PARP inhibitor–resistant BRCA -mutant ovarian cancerPublication Name: Science Translational MedicinePublication Date: 2023-06-21PMCID: PMC10758289PMID: 37343085DOI: 10.1126/scitranslmed.add7872
- Recent updates on 1,2,3-triazole-containing hybrids with in vivo therapeutic potential against cancers: A mini-reviewPublication Name: European Journal of Medicinal ChemistryPublication Date: 2023-05-05PMID: 36893627DOI: 10.1016/j.ejmech.2023.115254
- Development of BET Inhibitors as Potential Treatments for Cancer: Optimization of Pharmacokinetic PropertiesPublication Name: ACS Medicinal Chemistry LettersPublication Date: 2022-07-05PMCID: PMC9290009PMID: 35859878DOI: 10.1021/acsmedchemlett.2c00219
- Synthesis of BMS-986158Publication Name: SynfactsPublication Date: 2021-11-17DOI: 10.1055/s-0041-1737090
- Development of BET inhibitors as potential treatments for cancer: A new carboline chemotypePublication Name: Bioorganic & Medicinal Chemistry LettersPublication Date: 2021-11-01PMID: 34560263DOI: 10.1016/j.bmcl.2021.128376
- Discovery and Preclinical Pharmacology of an Oral Bromodomain and Extra-Terminal (BET) Inhibitor Using Scaffold-Hopping and Structure-Guided Drug DesignPublication Name: Journal of Medicinal ChemistryPublication Date: 2021-09-20PMID: 34543572DOI: 10.1021/acs.jmedchem.1c00625
- Retrospective assessment of rat liver microsomal stability at NCATS: data and QSAR modelsPublication Name: Scientific ReportsPublication Date: 2020-11-26PMCID: PMC7693334PMID: 33244000DOI: 10.1038/s41598-020-77327-0
- High-Throughput Screening to Identify Inhibitors of the Type I Interferon–Major Histocompatibility Complex Class I Pathway in Skeletal MusclePublication Name: ACS Chemical BiologyPublication Date: 2020-05-27PMCID: PMC7859889PMID: 32459468DOI: 10.1021/acschembio.0c00343
- Predictive models of aqueous solubility of organic compounds built on A large dataset of high integrityPublication Name: Bioorganic & Medicinal ChemistryPublication Date: 2019-07-15PMCID: PMC8274818PMID: 31176566DOI: 10.1016/j.bmc.2019.05.037
- Highly predictive and interpretable models for PAMPA permeabilityPublication Name: Bioorganic & Medicinal ChemistryPublication Date: 2017-02-01PMCID: PMC5291813PMID: 28082071DOI: 10.1016/j.bmc.2016.12.049
- BET inhibitor resistance emerges from leukaemia stem cellsPublication Name: NaturePublication Date: 2015-09-14PMCID: PMC6069604PMID: 26367796DOI: 10.1038/nature14888
- Efficacy of BET Bromodomain Inhibition in Kras-Mutant Non–Small Cell Lung CancerPublication Name: Clinical cancer research : an official journal of the American Association for Cancer ResearchPublication Date: 2013-11-14PMCID: PMC3838895PMID: 24045185DOI: 10.1158/1078-0432.ccr-12-3904
- Discovery and Preclinical Evaluation of [4-[[1-(3-fluorophenyl)methyl]-1H-indazol-5-ylamino]-5-methylpyrrolo[2,1-f][1,2,4]triazin-6-yl]carbamic Acid, (3S)-3-Morpholinylmethyl Ester (BMS-599626), a Selective and Orally Efficacious Inhibitor of Human Epidermal Growth Factor Receptor 1 and 2 KinasesPublication Name: Journal of Medicinal ChemistryPublication Date: 2009-10-12PMID: 19821562DOI: 10.1021/jm9010065
- [Statistical analysis of cerebrospinal fluid acid-base equilibrium and cerebrospinal fluid lactate concentration in cases of brain tumors, cerebrocranial injuries and meningoencephalitis]Publication Name: Neurologia i neurochirurgia polskaPublication Date: 1976-07PMID: 8740
PAT
- Tricyclic compounds as anticancer agentsPublication Number: WO-2015100282-A1Priority Date: 2013-12-24
- Tricyclic compound as anticancer agentsPublication Number: EP-3466949-B1Priority Date: 2013-12-24Grant Date: 2020-12-23
- Novel tricyclic compounds as anticancer agentsPublication Number: TW-202028203-APriority Date: 2013-12-24
- Novel tricyclic compounds as anticancer agentsPublication Number: TW-I726544-BPriority Date: 2013-12-24Grant Date: 2021-05-01
- Tricyclic compounds as anticancer agentsPublication Number: CN-108558871-BPriority Date: 2013-12-24Grant Date: 2022-02-18
AS ON JUNE2025 4.45 LAKHS VIEWS ON BLOG WORLDREACH AVAILABLEFOR YOUR ADVERTISEMENT
join me on Linkedin
Anthony Melvin Crasto Ph.D – India | LinkedIn
join me on Researchgate
RESEARCHGATE
join me on Facebook
Anthony Melvin Crasto Dr. | Facebook
join me on twitter
Anthony Melvin Crasto Dr. | twitter
+919321316780 call whatsaapp
EMAIL. amcrasto@gmail.com
……
| Clinical data | |
|---|---|
| Other names | BMS-986158 |
| Identifiers | |
| IUPAC name | |
| CAS Number | 1800340-40-2 |
| PubChem CID | 118196485 |
| DrugBank | DB15435 |
| ChemSpider | 58828664 |
| UNII | X8BW0MQ5PI |
| KEGG | D12710 |
| ChEMBL | ChEMBL4297458 |
| Chemical and physical data | |
| Formula | C30H33N5O2 |
| Molar mass | 495.627 g·mol−1 |
| 3D model (JSmol) | Interactive image |
| SMILES | |
| InChI | |
References
- Ma Z, Zhang C, Bolinger AA, Zhou J (October 2024). “An updated patent review of BRD4 degraders”. Expert Opinion on Therapeutic Patents. 34 (10): 929–951. doi:10.1080/13543776.2024.2400166. PMC 11427152. PMID 39219068.
- “Clinical Trials Using Ezobresib”. National Cancer Institute.
- Brown A. “Bristol backs out of BET inhibition”. ApexOnco.
////////////Ezobresib, antineoplastic, BMS-986158, BMS 986158, Bristol Myers Squibb, antineoplastic, UNII-X8BW0MQ5PI
Imlunestrant
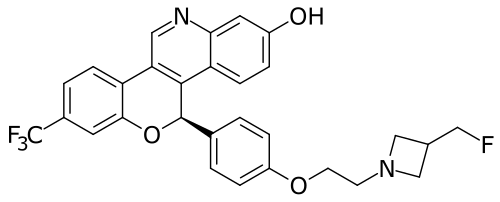
Imlunestrant
CAS 2408840-26-4
as tosylate: 2408840-41-3
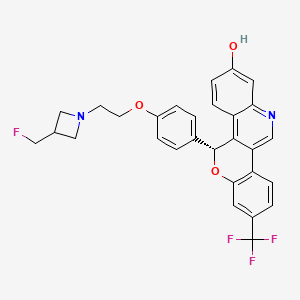
(5R)-5-[4-[2-[3-(fluoromethyl)azetidin-1-yl]ethoxy]phenyl]-8-(trifluoromethyl)-5H-chromeno[4,3-c]quinolin-2-ol
- (5r)-5-(4-(2-(3-(fluoromethyl)azetidin-1-yl)ethoxy)phenyl)-8-(trifluoromethyl)-5h-(1)benzopyrano(4,3-c)quinolin-2-ol
- 5h-(1)benzopyrano(4,3-c)quinolin-2-ol, 5-(4-(2-(3-(fluoromethyl)-1-azetidinyl)ethoxy)phenyl)-8-(trifluoromethyl)-, (5r)-
MF C29H24F4N2O3 MW 524.516
FDA 9/25/2025, Inluriyo, LY3484356, LY-3484356, To treat estrogen receptor-positive, human epidermal growth factor receptor 2-negative, estrogen receptor-1-mutated advanced or metastatic breast cancer with disease progression following at least one line of endocrine therapy
Imlunestrant, sold under the brand name Inluriyo, is an anti-cancer medication used for the treatment of breast cancer.[1] It is an is an estrogen receptor antagonist.[1] It is used as the salt, imlunestrant tosylate.[2] It is taken by mouth.[1] It was developed by Eli Lilly and Company.[2]
The most common adverse events and laboratory abnormalities include decreased hemoglobin, musculoskeletal pain, decreased calcium, decreased neutrophils, increased AST, fatigue, diarrhea, increased ALT, increased triglycerides, nausea, decreased platelets, constipation, increased cholesterol, and abdominal pain.[2]
Imlunestrant was approved for medical use in the United States in September 2025.[2]
SYN
- Imlunestrant with or without Abemaciclib in Advanced Breast CancerPublication Name: The New England journal of medicinePublication Date: 2025-03-27PMID: 39660834DOI: 10.1056/nejmoa2410858
- Targeting the Estrogen Receptor for the Treatment of Breast Cancer: Recent Advances and ChallengesPublication Name: Journal of Medicinal ChemistryPublication Date: 2023-06-28PMID: 37377342DOI: 10.1021/acs.jmedchem.3c00136
- Novel endocrine therapies: What is next in estrogen receptor positive, HER2 negative breast cancer?Publication Name: Cancer Treatment ReviewsPublication Date: 2023-06PMID: 37146385DOI: 10.1016/j.ctrv.2023.102569
- Oral Selective Estrogen Receptor Degraders (SERDs) as a Novel Breast Cancer Therapy: Present and Future from a Clinical PerspectivePublication Name: International Journal of Molecular SciencesPublication Date: 2021-07-22PMCID: PMC8345926PMID: 34360578DOI: 10.3390/ijms22157812
PAT
https://patentscope.wipo.int/search/en/detail.jsf?docId=US281655517&_cid=P12-MG7DCV-14904-1
Example 1A
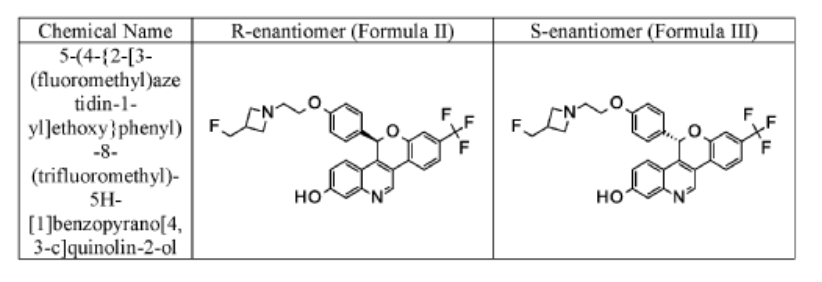
5-(4-{2-[3-(Fluoromethyl)azetidin-1-yl]ethoxy}phenyl)-8-(trifluoromethyl)-5H-[1]benzopyrano[4,3-c]quinolin-2-ol, Isomer 1Separate the two enantiomers of 5-(4-{2-[3-(fluoromethyl)azetidin-1-yl]ethoxy}phenyl)-8-(trifluoromethyl)-5H-[1]benzopyrano[4,3-c]quinolin-2-ol by chiral SFC with the following conditions: Column: LUX® Cellulose-1, 5×25 cm; eluting with a mobile phase of 30% iPrOH (with 0.5% DMEA) in CO 2; column temperature: 40° C.; flow rate: 300 g/minute; UV detection wavelength: 270 nm to give Example 1A as the first eluting enantiomer (Isomer 1). ES/MS (m/z): 525.2 (M+H). Confirm enantiomeric enrichment of Isomer 1 by chiral analytical SFC, >99% ee, t (R): 1.30 minutes; column: CHIRALCEL® OD-H, 4.6×150 mm; eluting with a mobile phase of 30% MeOH (0.2% IPA) in CO 2; column temperature: 40° C.; flow rate: 5 mL/minute; UV detection wavelength: 225 nm. Isolate the title compound of Example 1B to give the second eluting enantiomer (Isomer 2). ES/MS (m/z): 525.2 (M+H). Confirm enantiomeric enrichment of Isomer 2 by chiral analytical SFC, 98% ee, t (R): 2.03 minutes; column: CHIRALCEL® OD-H, 4.6×150 mm; eluting with a mobile phase of 30% MeOH (0.2% IPA) in CO 2; column temperature: 40° C.; flow rate: 5 mL/minute; UV detection wavelength: 225 nm.
Alternate Preparation Example 1B
Crystalline 5-(4-{2-[3-(Fluoromethyl)azetidin-1-yl]ethoxy}phenyl)-8-(trifluoromethyl)-5H-[1]benzopyrano[4,3-c]quinolin-2-ol, Isomer 2
PAT
https://patentscope.wipo.int/search/en/detail.jsf?docId=WO2020014435&_cid=P12-MG7DHN-18354-1
EXAMPLE 1
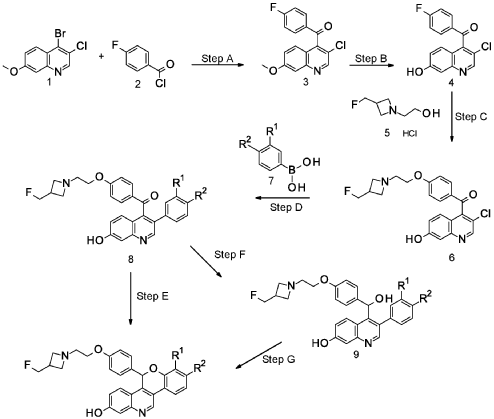
Racemic 5-(4-{2-[3-(Fluoromethyl)azetidin-l-yl]ethoxy}phenyl)-8-(trifluoromethyl)-5H- [ 1 ]benzopyrano[4,3 -c]quinolin-2-ol
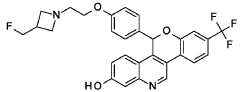
Cool a solution of (4-{2-[3-(fluoromethyl)azetidin-l-yl]ethoxy}phenyl){3-[2-fluoro-4-(trifluoromethyl)phenyl]-7-hydroxyquinolin-4-yl}methanone (5.27 g, 9.71 mmol) in 1,4-dioxane (100 mL) to 5 °C. Add lithium triethylborohydride (1 M in THF, 30.0 mL, 30.0 mmol). Remove the cooling bath and stir for 1.5 hours at room temperature. Quench the mixture with water. Add saturated NH4Cl solution and EtOAc. Separate the layers and extract the aqueous layer with EtOAc. Combine the organic extracts, dry over anhydrous MgS04, filter, and concentrate the filtrate. Dissolve the crude residue in THF (100 mL).
Add sodium hydride (60% in mineral oil, 1.94 g, 48.5 mmol). Reflux the solution for 1.5 hours. Add additional sodium hydride (60% in mineral oil, 1.94 g, 48.5 mmol), then reflux for an additional 30 minutes. Cool the solution to room temperature and quench with water. Add EtOAc and saturated NH4Cl solution. Separate the layers and extract the aqueous layer with EtOAc. Combine the organic extract, dry over anhydrous MgS04, filter, and concentrate the filtrate. Purify the residue by silica gel column chromatography eluting with a gradient of 5-7% MeOH in DCM to give the title compound (3.70 g, 72%) as a light yellow foam. ES/MS (m/z): 525.2 (M+H).
Prepare the following compounds in a manner essentially analogous to the method of Example 1, with the following variations in procedure. For the reduction, use 3 to 5 equivalents of lithium triethylborohydride with reaction times from 30 minutes to one hour and drying of the organic layers over magnesium sulfate or sodium sulfate. ETse the crude residue directly or purify by silica gel column chromatography eluting with a gradient of 0-5-7.5-10% MeOH in DCM before cyclization. Complete the cyclization by refluxing in THF for up to 16 hours, or in DMF, from 2 hours at room temperature for Ex 2, to 2 hours at 85 °C for Ex 8. Extract with DCM or EtOAc and dry organic layers over magnesium sulfate or sodium sulfate. Purify by silica gel column chromatography using up to 10% (MeOH or 7 M ammoniated MeOH) in DCM (Ex 2: gradient 0-10% MeOH in DCM; Ex 5: gradient 4-10% 7 M ammoniated MeOH in DCM; Ex 8: gradient 5-7.5% 7 M ammoniated MeOH in DCM) or by high pH reversed phase HPLC as noted.
EXAMPLE 1A
-(4-{2-[3-(Fluoromethyl)azetidin-l-yl]ethoxy}phenyl)-8-(trifluoromethyl)-5H- [l]benzopyrano[4,3-c]quinolin-2-ol, Isomer 1
and
EXAMPLE 1B
5-(4-{2-[3-(Fluoromethyl)azetidin-l-yl]ethoxy}phenyl)-8-(trifluoromethyl)-5H- [l]benzopyrano[4,3-c]quinolin-2-ol, Isomer 2
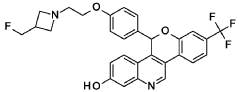
Separate the two enantiomers of 5-(4-{2-[3-(fluoromethyl)azetidin-l-yl]ethoxy}phenyl)-8-(trifluoromethyl)-5H-[l]benzopyrano[4,3-c]quinolin-2-ol by chiral SFC with the following conditions: Column: LUX® Cellulose-l, 5 x 25 cm; eluting with a mobile phase of 30% iPrOH (with 0.5% DMEA) in C02; column temperature: 40 °C; flow rate: 300 g/minute; UV detection wavelength: 270 nm to give Example 1 A as the first eluting enantiomer (Isomer 1). ES/MS (m/z): 525.2 (M+H). Confirm enantiomeric enrichment of Isomer 1 by chiral analytical SFC, >99% ee, /(R>: 1.30 minutes; column: CHFRALCEL® OD-H, 4.6 x 150 mm; eluting with a mobile phase of 30% MeOH (0.2% IP A) in C02; column temperature: 40 °C; flow rate: 5 mL/minute; UV detection wavelength: 225 nm. Isolate the title compound of Example 1B to give the second eluting enantiomer (Isomer 2). ES/MS (m/z): 525.2 (M+H). Confirm enantiomeric enrichment of Isomer 2 by chiral analytical SFC, 98% ee, /(R>: 2.03 minutes; column: CHIRALCEL® OD-H, 4.6 x 150 mm; eluting with a mobile phase of 30% MeOH (0.2% IP A) in C02; column temperature: 40 °C; flow rate: 5 mL/minute; UV detection wavelength: 225 nm.
Alternate Preparation EXAMPLE 1B
Crystalline 5-(4-{2-[3-(Fluoromethyl)azetidin-l-yl]ethoxy}phenyl)-8-(trifluoromethyl)-5H- [l]benzopyrano[4,3-c]quinolin-2-ol, Isomer 2
Stir 5-(4-{2-[3-(fluoromethyl)azetidin-l-yl]ethoxy}phenyl)-8-(trifluoromethyl)-5H-[l]benzopyrano[4,3-c]quinolin-2-ol, 4-methylbenzenesulfonic acid, Isomer 2 (23.8 g, 0.034 mol) in water (250 mL) at 1000 rpm. Add NaOH (76 pL) and stir the solution for 2 hours. Add DCM (600 mL). Separate the mixture, dry the DCM extract with magnesium sulfate, filter the material through a syringe filter (0.45 pm), and concentrate to dryness. Allow the material to sit under a N2 stream over a weekend. Add 1 : 1 EtOH/water (80 mL) and stir the mixture with sonication. Collect a tan solid by filtration on a nylon membrane to give the title compound (10.47 g, 0.02 mol, 59%).
PAT
- Selective estrogen receptor degradersPublication Number: US-2023234960-A1Priority Date: 2018-07-12
- Selective estrogen receptor degraderPublication Number: CN-112424205-BPriority Date: 2018-07-12Grant Date: 2023-10-31
- selective estrogen receptor degraderPublication Number: CN-117379428-APriority Date: 2018-07-12
- Selective estrogen receptor degradersPublication Number: US-11993608-B2Priority Date: 2018-07-12Grant Date: 2024-05-28
- Selective estrogen receptor degradersPublication Number: US-12128040-B2Priority Date: 2018-07-12Grant Date: 2024-10-29
PAT
https://patents.google.com/patent/US11926634B2/en
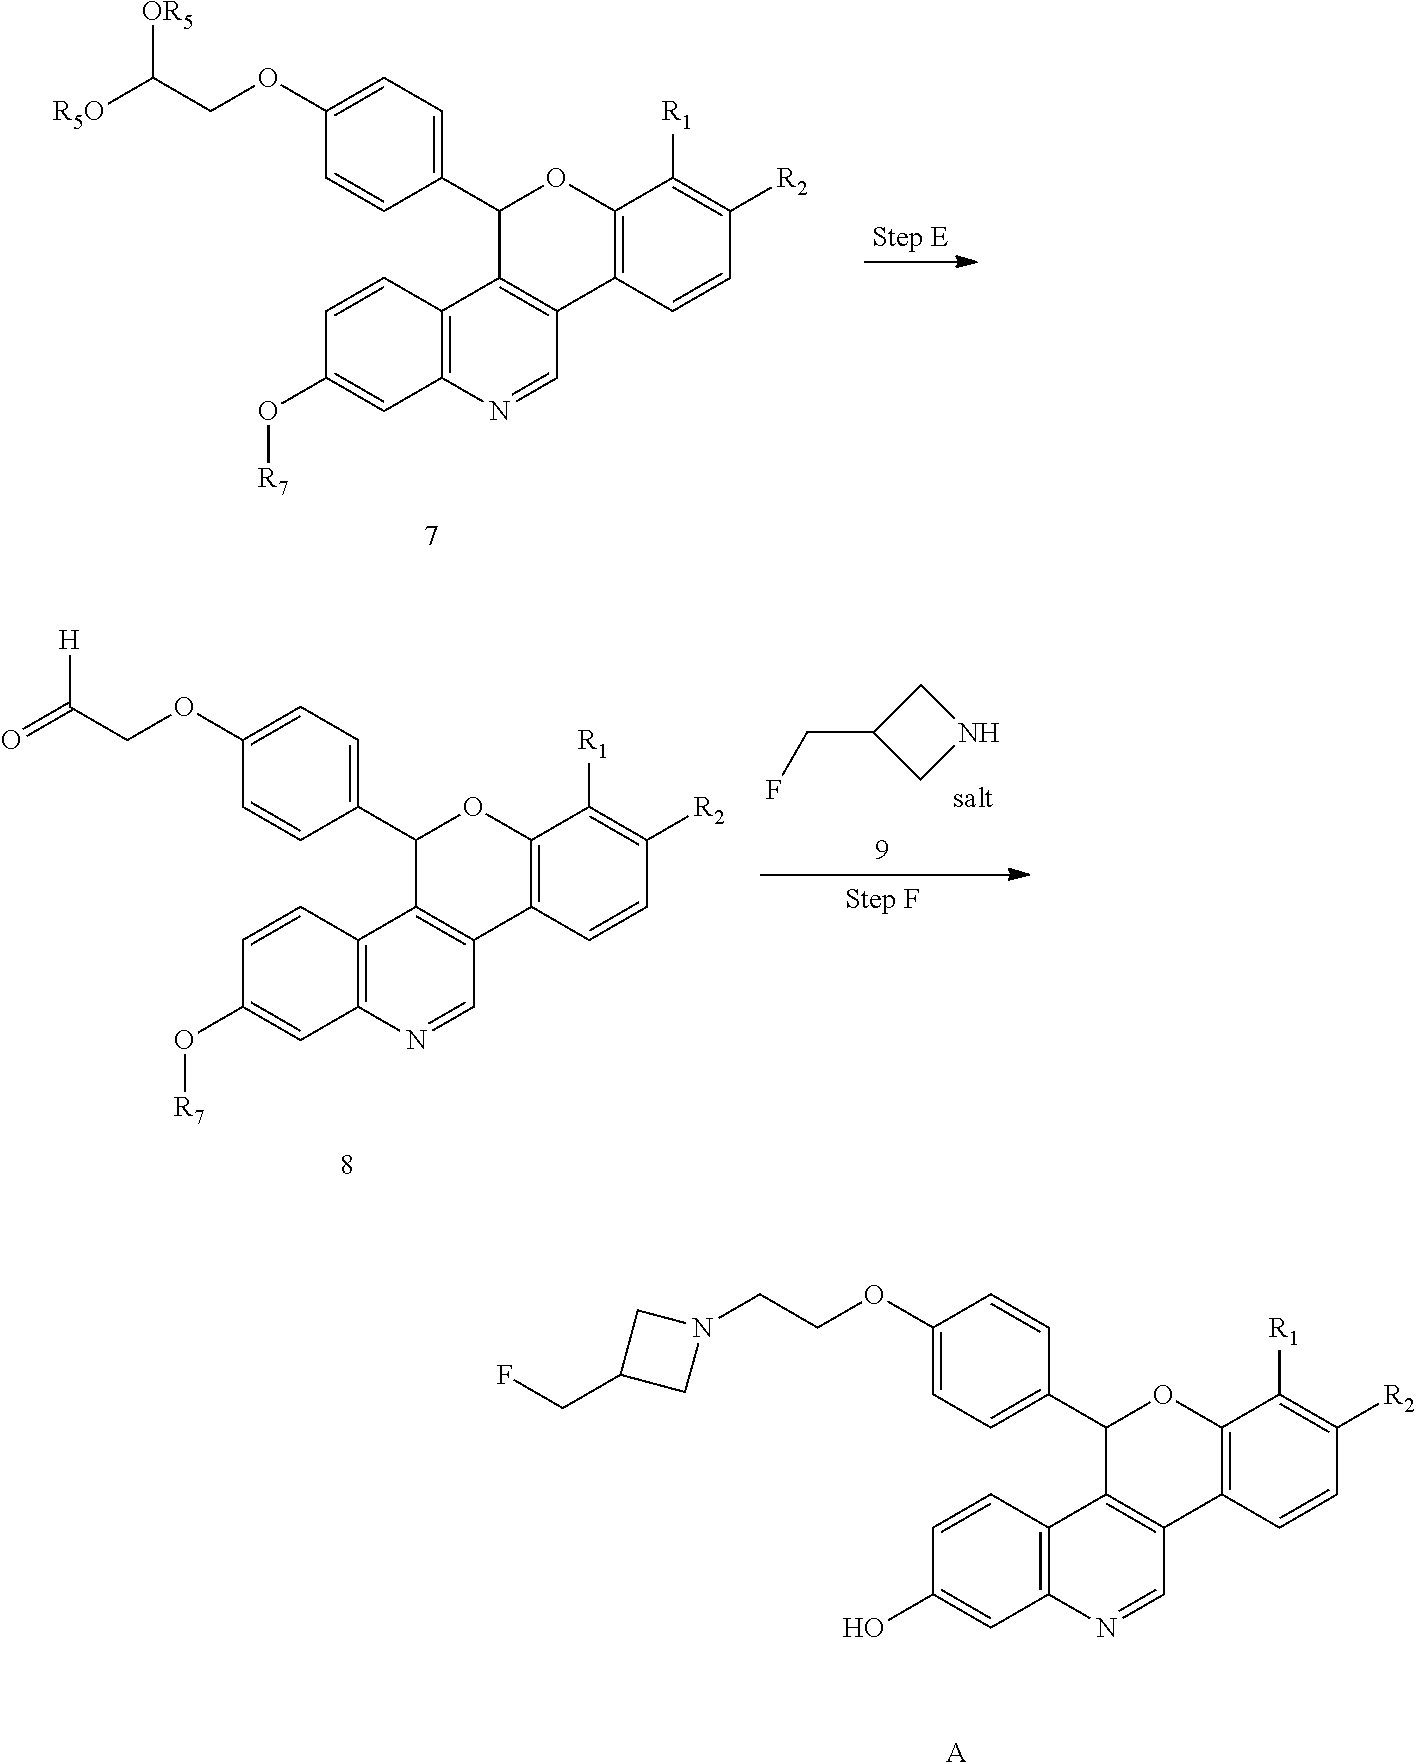
Selective estrogen receptor degraders (SERDs) bind to the estrogen receptor (ER) and downregulate ER-mediated transcriptional activity. The degradation and downregulation caused by SERDs can be useful in the treatment of various proliferative immune mediated disorders, cell proliferation disorders, including cancers such as breast cancer, ovarian cancer, endometrial cancer, prostate cancer, uterine cancer, gastric cancer, and lung cancer as well as mutations due to emerging resistance. Some small molecule examples of SERDs have been disclosed in the literature (see, e.g., WO2005073204, WO2014205136, and WO2016097071). Nonetheless, there is a need for new SERDs to treat ER-positive cancers, such as breast cancer, gastric cancer, and/or lung cancer.
As described in U.S. Pat. No. 10,654,866 (the ‘866 patent) a series of SERDs of the following formula have been discovered, along with pharmaceutically acceptable salts thereof:
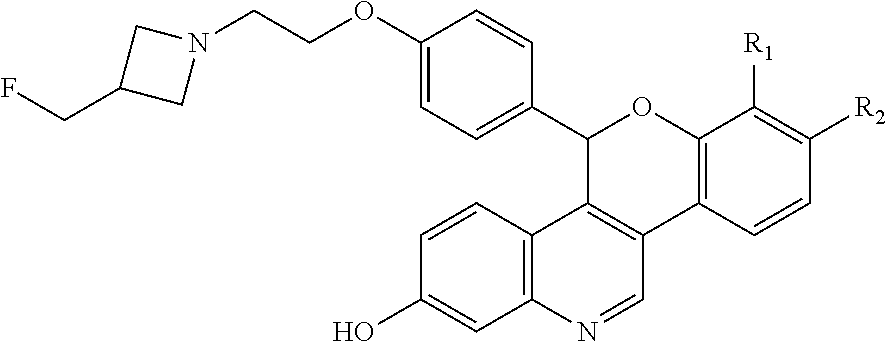
wherein one of R1 and R2 are independently Cl, F, —CF3, or —CH3, and the other is H.
AS ON JUNE2025 4.45 LAKHS VIEWS ON BLOG WORLDREACH AVAILABLEFOR YOUR ADVERTISEMENT
join me on Linkedin
Anthony Melvin Crasto Ph.D – India | LinkedIn
join me on Researchgate
RESEARCHGATE
join me on Facebook
Anthony Melvin Crasto Dr. | Facebook
join me on twitter
Anthony Melvin Crasto Dr. | twitter
+919321316780 call whatsaapp
EMAIL. amcrasto@gmail.com
……
| Clinical data | |
|---|---|
| Trade names | Inluriyo |
| Other names | LY3484356, LY-3484356 |
| AHFS/Drugs.com | Inluriyo |
| License data | US DailyMed: Imlunestrant |
| Routes of administration | By mouth |
| Drug class | Estrogen receptor antagonist |
| ATC code | None |
| Legal status | |
| Legal status | US: ℞-only[1] |
| Identifiers | |
| IUPAC name | |
| CAS Number | 2408840-26-4as tosylate: 2408840-41-3 |
| PubChem CID | 146603228 |
| DrugBank | DB19043 |
| ChemSpider | 115010421 |
| UNII | 9CXQ3PF69Uas tosylate: F7UDT90EW5 |
| KEGG | D12216as tosylate: D12217 |
| ChEMBL | ChEMBL5095183 |
| Chemical and physical data | |
| Formula | C29H24F4N2O3 |
| Molar mass | 524.516 g·mol−1 |
| 3D model (JSmol) | Interactive image |
| SMILES | |
| InChI | |
References
- https://www.accessdata.fda.gov/drugsatfda_docs/label/2025/218881s000lbl.pdf
- “FDA approves imlunestrant for ER-positive, HER2-negative, ESR1-mutated advanced or metastatic breast cancer”. U.S. Food and Drug Administration (FDA). 25 September 2025. Retrieved 27 September 2025. This article incorporates text from this source, which is in the public domain.
- “U.S. FDA approves Inluriyo (imlunestrant) for adults with ER+, HER2-, ESR1-mutated advanced or metastatic breast cancer” (Press release). Eli Lilly. 25 September 2025. Retrieved 27 September 2025 – via PR Newswire.
- World Health Organization (2022). “International nonproprietary names for pharmaceutical substances (INN): recommended INN: list 88”. WHO Drug Information. 36 (3). hdl:10665/363551.
Further reading
- Jhaveri, Komal L.; Jeselsohn, Rinath; Lim, Elgene; Hamilton, Erika P.; Yonemori, Kan; Beck, J. Thaddeus; et al. (June 2022). “A phase 1a/b trial of imlunestrant (LY3484356), an oral selective estrogen receptor degrader (SERD) in ER-positive (ER+) advanced breast cancer (aBC) and endometrial endometrioid cancer (EEC): Monotherapy results from EMBER”. Journal of Clinical Oncology. 40 (16_suppl): 1021. doi:10.1200/JCO.2022.40.16_suppl.1021. S2CID 249445691.
- Jhaveri, Komal; O’Shaughnessy, Joyce; Andre, Fabrice; Goetz, Matthew P.; Harbeck, Nadia; Martín, Miguel; et al. (March 2023). “Abstract OT1-01-02: EMBER-4: A phase 3 adjuvant trial of imlunestrant vs standard endocrine therapy (ET) in patients with ER+, HER2- early breast cancer (EBC) with an increased risk of recurrence who have previously received 2 to 5 years of adjuvant ET”. Cancer Research. 83 (5_Supplement): OT1–01–02-OT1-01–02. doi:10.1158/1538-7445.SABCS22-OT1-01-02.
- Neven, P.; Stahl, N.; Losada, M.J. Vidal; Jimenez, M. Martin; Kaufman, P.A.; Harbeck, N.; et al. (October 2023). “273P A preoperative window-of-opportunity (WOO) study of imlunestrant in ER+, HER2- early breast cancer (EBC): Final analysis from EMBER-2”. Annals of Oncology. 34: S292 – S293. doi:10.1016/j.annonc.2023.09.470. S2CID 264385454.
External links
- Clinical trial number NCT04975308 for “A Study of Imlunestrant, Investigator’s Choice of Endocrine Therapy, and Imlunestrant Plus Abemaciclib in Participants With ER+, HER2- Advanced Breast Cancer (EMBER-3)” at ClinicalTrials.gov
/////////Imlunestrant, FDA 2025, APPROVALS 2025, Inluriyo, CANCER, LY3484356, LY 3484356, 9CXQ3PF69U
Oritinib
Oritinib
- CAS 2035089-28-0
- MESYLATE CAS 2180164-79-6
- SH-1028
- SK593H37SC
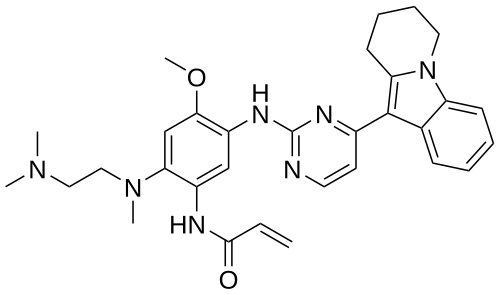
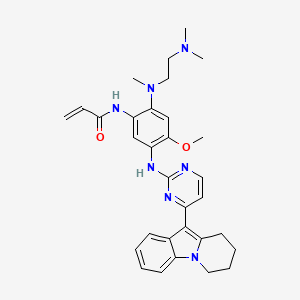
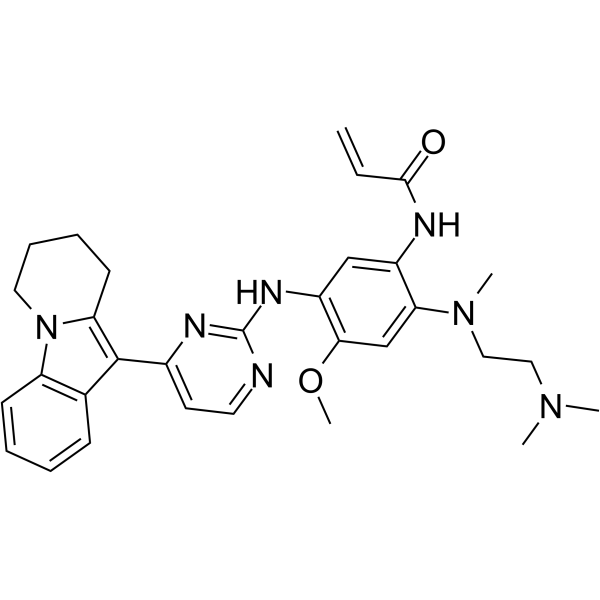
- N-[2-[2-(dimethylamino)ethyl-methylamino]-4-methoxy-5-[[4-(6,7,8,9-tetrahydropyrido[1,2-a]indol-10-yl)pyrimidin-2-yl]amino]phenyl]prop-2-enamide
- 539.7 g/mol, C31H37N7O2
- rilertinib
CHINA 2024, Nanjing Sanhome Pharmaceutical.
N-[2-[2-(dimethylamino)ethyl-methylamino]-4-methoxy-5-[[4-(6,7,8,9-tetrahydropyrido[1,2-a]indol-10-yl)pyrimidin-2-yl]amino]phenyl]prop-2-enamide
Oritinib is an investigational new drug currently under investigation for its potential use in cancer treatment.[1][2] As a epidermal growth factor receptor (EGFR) tyrosine kinase inhibitor, oritinib targets specific enzymes involved in the signaling pathways that regulate cell division and survival, which are often dysregulated in cancer cells.[1]
Oritinib (SH-1028), an irreversible third-generation EGFR TKI, overcomes T790M-mediated resistance in non-small cell lung cancer. Oritinib (SH-1028), a mutant-selective inhibitor of EGFR kinase activity, inhibits EGFRWT, EGFRL858R, EGFRL861Q, EGFRL858R/T790M, EGFRd746-750 and EGFRd746-750/T790M kinases, with IC50s of 18, 0.7, 4, 0.1, 1.4 and 0.89 nM, respectively.
PAT
https://patents.google.com/patent/CN115974845B/en
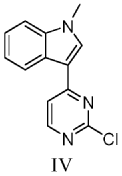
Reaction condition optimization experiment:
The experimental group numbered 1 referred to in table 1 below is the preparation of 1-methyl-3- (2-chloro-4-pyrimidinyl) indole, which was prepared as follows:
To a 10mL reaction tube, 2, 4-dichloropyrimidine (74.5 mg,0.05 mol), zinc triflate (67.3 mg,0.37 equiv), scandium triflate (7.4 mg,0.03 equiv) and 1-methylindole (78.6 mg,1.2 equiv) were added under inert gas atmosphere, and acetonitrile (2.5 mL) were heated to 80℃to react for 24 hours. The reaction was quenched with 30ml of ethyl acetate, the above mixture was added to a separating funnel, 50ml of saturated aqueous sodium carbonate and 50ml of saturated aqueous ammonium chloride were added thereto, and the mixture was shaken for 2 minutes, and the organic phase was taken after the liquid in the separating funnel had settled and separated. The aqueous phase was rinsed with 30ml of ethyl acetate under shaking for 2 times, the whole organic phase was collected, silica gel powder and anhydrous sodium sulfate were added thereto, and the mixture was dried under reduced pressure and packed into a silica gel column. Sequential gradient elution was performed using 250ml (PE: EA: triethylamine 16:4:1), 250ml (PE: EA: triethylamine 15:5:1), 250ml (PE: EA: triethylamine 40:20:3) as developing reagent. The eluent is collected and dried under reduced pressure to obtain pale yellow solid with the yield of 90 percent.
The nuclear magnetic resonance spectrum of 1-methyl-3- (2-chloro-4-pyrimidinyl) indole is as follows:
1H NMR(400MHz,DMSO-d6)δ8.51(d,J=5.9Hz,2H),8.40(dd,1H),7.82(d,J=5.4Hz,1H),7.56(dd,1H),7.28(pd,J=7.1,1.4Hz,2H),3.88(s,3H).
13C NMR(101MHz,DMSO)δ164.55,160.32,158.75,137.84,134.83,125.30,122.81,121.74,121.64,114.43,110.90,110.76,33.31.
PAT
CN109705118
https://patentscope.wipo.int/search/en/detail.jsf?docId=CN242181067&_cid=P20-MEGI3F-20821-1
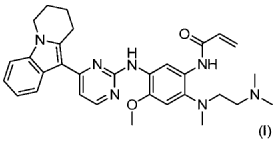
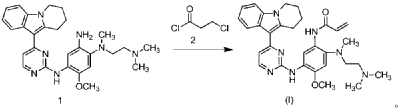
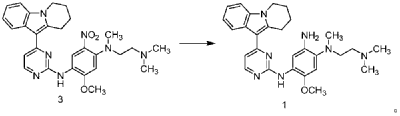
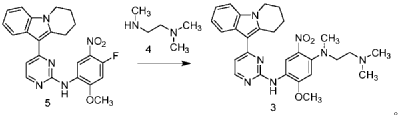
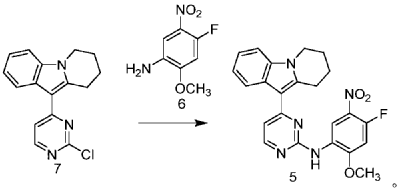
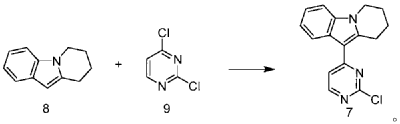
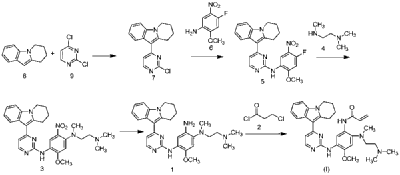
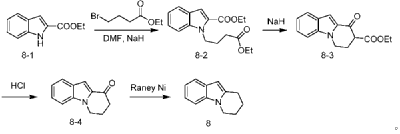
| Step 1: Synthesis of 10-(2-chloropyrimidin-4-yl)-6,7,8,9-tetrahydropyrido[1,2-a]indole |
| |
| In a 100L vertical jacketed glass reactor, add ethylene glycol dimethyl ether (39.15kg) and 2,4-dichloropyrimidine (3.915kg). Cool the solid-liquid mixture to below 10°C, then add anhydrous aluminum chloride (3.855kg) in batches, controlling the addition rate to keep the temperature below 30°C. After the addition is complete, stir at 25±5°C for 30 minutes, then add 6,7,8,9-tetrahydropyrido[1,2-a]indole (4.500kg). Raise the temperature to 60±5°C and react for 3 hours. Monitor by HPLC until the 6,7,8,9-tetrahydropyrido[1,2-a]indole content does not exceed 1.0%, confirming the reaction is complete. The reaction solution was cooled to below 25° C., purified water (90.0 kg) was added, stirred, and filtered. The filter cake was added to acetonitrile (17.8 kg), slurried, filtered, and dried to obtain a yellow powdery solid, a total of 6.652 kg, with a yield of 89.2%. |
| Step 2: Synthesis of N-(4-fluoro-2-methoxy-5-nitrophenyl)-4-(6,7,8,9-tetrahydropyrido[1,2-a]indol-10-yl)pyrimidin-2-amine |
| |
| To a 500L glass-lined reactor, sec-butyl alcohol (80.82kg), 10-(2-chloropyrimidin-4-yl)-6,7,8,9-tetrahydropyrido[1,2-a]indole (6.652kg), 4-fluoro-2-methoxy-5-nitroaniline (4.363kg), and p-toluenesulfonic acid monohydrate (4.816kg) were added to obtain a solid-liquid mixture. The reaction mixture was heated to reflux, and the solid gradually dissolved. As the reaction proceeded, a yellow solid precipitated. After reflux for 7.5 hours, the reaction was monitored by HPLC to confirm completion. Heating was stopped, the reaction mixture was cooled to below 15°C, stirred for 1 hour, and the solid was centrifuged and filtered. Acetonitrile (31.5kg) was added to the filter cake, and the mixture was slurried at 25±5°C for 1.5 hours. The mixture was centrifuged and dried to obtain the title compound, a total of 9.548kg, with a yield of 94.0%. |
| Step 3: Synthesis of N 1 -(2-dimethylaminoethyl)-5-methoxy-N 1 -methyl-2-nitro-N 4 -(4-(6,7,8,9-tetrahydropyrido[1,2-a]indol-10-yl)pyrimidin-2-yl)phenyl-1,4-diamine |
| |
| To a 100 L vertical jacketed glass reactor, add N,N-dimethylacetamide (44.7 kg), N-(4-fluoro-2-methoxy-5-nitrophenyl)-4-(6,7,8,9-tetrahydropyrido[1,2-a]indol-10-yl)pyrimidin-2-amine (9.548 kg), N,N,N’-trimethylethylenediamine (3.380 kg), and N,N-diisopropylethylamine (4.841 kg). Under nitrogen, the reaction mixture was reacted at 85±5°C for 2 hours and monitored by HPLC until the reaction was complete. The reaction solution was cooled to below 70°C, purified water (95.5 kg) was added, filtered, and dried to obtain the title compound, a total of 8.206 kg, with a yield of 72.2%. |
| Step 4: Synthesis of N 1 -(2-(dimethylamino)ethyl)-5-methoxy-N 1 -methyl-N 4 -(4-(6,7,8,9-tetrahydropyrido[1,2-a]indol-10-yl)pyrimidin-2-yl)benzene-1,2,4-triamine |
| |
| A 100 L vertical jacketed reactor was charged with anhydrous ethanol (32.39 kg), purified water (14.32 kg), N 1 -(2-dimethylaminoethyl)-5-methoxy-N 1 -methyl-2-nitro-N 4 -(4-(6,7,8,9-tetrahydropyrido[1,2-a]indol-10-yl)pyrimidin-2-yl)phenyl-1,4-diamine (4.103 kg), reduced iron powder (2.224 kg), and ammonium chloride (2.129 kg). The reaction mixture was refluxed for 1.5 hours and monitored by HPLC until the reaction was complete. The reaction mixture was cooled to below 50°C and filtered through diatomaceous earth to remove the solid. The filtrate was concentrated, and tetrahydrofuran (3.45 kg) and purified water (34.71 kg) were added to the residue. The mixture was slurried, filtered, and dried to obtain 3.244 kg of the title compound in an 84.0% yield. |
| Step 5: Synthesis of N-(2-((2-(dimethylamino)ethyl)(methyl)amino)-4-methoxy-5-((4-(6,7,8,9-tetrahydropyrido[1,2-a]indol-10-yl)pyrimidin-2-yl)amino)phenyl)allylamide |
| |
| Add N,N-dimethylacetamide (48.6 kg) to a 100 L vertical jacketed glass reactor. Raise the temperature to 40°C, then add N₁- ( 2-(dimethylamino)ethyl)-5-methoxy- N₁ -methyl- N₄- (4-(6,7,8,9-tetrahydropyrido[1,2-a]indol-10-yl)pyrimidin-2-yl)benzene-1,2,4-triamine (6.487 kg). Then, begin the dropwise addition of 3-chloropropionyl chloride (1.777 kg). Control the addition rate to no more than 60°C. After the addition is complete, cool the reaction mixture and stir at 40±5°C for 1 hour. Sample the mixture and monitor the reaction by HPLC until complete. Add purified water (0.253 kg) and stir for 30 minutes. |
| The reaction mixture was heated at 80±5°C, triethylamine (13.52 kg) was added, and the temperature was raised to 95±5°C. After reacting for 2 hours, the reaction was complete as determined by HPLC. The temperature was then lowered, and methanol (83.0 kg) was added. The mixture was then cooled and crystallized, filtered, and dried to obtain 4.953 kg of the title compound, with a yield of 68.6% and a purity of 97.37%. |
| Step 6: Purification of N-(2-((2-(dimethylamino)ethyl)(methyl)amino)-4-methoxy-5-((4-(6,7,8,9-tetrahydropyrido[1,2-a]indol-10-yl)pyrimidin-2-yl)amino)phenyl)allylamide |
| Anhydrous ethanol (31.25 kg) was added to a 100 L reactor and heated to above 70°C. The crude N-(2-((2-(dimethylamino)ethyl)(methyl)amino)-4-methoxy-5-((4-(6,7,8,9-tetrahydropyrido[1,2-a]indol-10-yl)pyrimidin-2-yl)amino)phenyl)allylamide prepared in step 5 was added. The reaction mixture was heated and stirred under nitrogen until dissolved. The reaction mixture was cooled to below 10°C, the precipitated solid was centrifuged and dried under vacuum at 60±5°C for more than 12 hours to obtain 4.559 kg of the title compound with a yield of 92.1% and a purity of 98.73%. 1 H NMR (300 MHz, DMSO-d 6 )δ10.20(s,1H),8.65(s,1H),8.34(d,1H),8.11(s,1H),8.06(d,1H),7.43(d, 1H),7.19-7.03(m,3H),6.98(s,1H),6.57-6.41(m,1H),6.28-6.15(m,1H),5.8 2-5.71(m,1H),4.09(t,2H),3.84(s,3H),3.18(t,2H),3.06-2.92(m,2H),2.66 (s,3H),2.47-2.40(m,2H),2.27(s,6H),2.08-1.96(m,2H),1.87-1.74(m,2H). ESI-Ms m/z: 540.3 [M+H] + . |
| Example 2: Synthesis of N-(2-((2-(dimethylamino)ethyl)(methyl)amino)-4-methoxy-5-((4-(6,7,8,9-tetrahydropyrido[1,2-a]indol-10-yl)pyrimidin-2-yl)amino)phenyl)allylamide |
| |
| The preparation method is the same as that in step 5 of Example 1, except that N,N-dimethylacetamide is replaced by N,N-dimethylformamide. The purity of the obtained title compound is 69%. |
| The N-(2-((2-(dimethylamino)ethyl)(methyl)amino)-4-methoxy-5-((4-(6,7,8,9-tetrahydropyrido[1,2-a]indol-10-yl)pyrimidin-2-yl)amino)phenyl)allylamide of the present invention prepared according to the above method has a high yield and purity, mild reaction conditions, easy purification, stable process, easy operation, environmental friendliness, and can meet the requirements of industrial-scale production and application. |
Syn
European Journal of Medicinal Chemistry 291 (2025) 117643
Oritinib represents a third-generation EGFR TKI engineered by Nanjing Sanhome Pharmaceutical. This agent specifically targets both EGFR-sensitizing mutations and the T790 M resistance mutation,
thereby addressing resistance mechanisms linked to prior-generation EGFR-TKIs. In 2024, the NMPA granted approval for Oritinib to treat adult patients with locally advanced or metastatic NSCLC who have experienced disease progression during or following EGFR-TKI therapy and possess confirmed EGFR T790 M mutation-positive status. The mechanism of action of Oritinib involves irreversible binding to mutant EGFR, including the T790 M variant, which in turn suppresses down stream signaling pathways responsible for tumor cell proliferation and survival [28]. The mechanism of Oritinib effectively inhibits tumor growth in patients harboring T790M-mediated resistance to first- and second-generation EGFR-TKIs. Clinical efficacy was established in a Phase II trial (NCT03823807) enrolling patients with EGFR T790 Mmutation-positive NSCLC who had experienced disease progression following prior EGFR-TKI therapy. This study documented an ORR of 60.5 % and a median PFS of 9.6 months, highlighting substantial anti
tumor efficacy in this specific patient cohort. In terms of safety, Oritinib exhibited favorable tolerability. The predominant treatment-related adverse events were rash, diarrhea, and elevated liver enzymes, pri
marily of mild (Grade 1) or moderate (Grade 2) severity. No dose-limiting toxicities were encountered, and the overall safety profile aligned with those observed for other third-generation EGFR-TKIs [29].
The synthetic route of Oritinib Mesylate, shown in Scheme 7, begins with nucleophilic substitution reaction between Orit-001 and Orit-002 to yield Orit-003, which further reacts with Orit-004 via nucleophilic substitution to produce Orit-005 [30]. Orit-005 subsequently undergoes another nucleophilic substitution with Orit-006 to generate Orit-007. Following this, Orit-007 is reduced to form Orit-008. Finally, an amidation reaction between Orit-008 and Orit-009 affords Oritinib.
[28] C. Zhou, A. Xiong, L. Miao, J. Chen, K. Li, H. Liu, Z. Ma, H. Wang, Z. Lu, J. Shen,
P51.03 oritinib (SH-1028), a third-generation EGFR-TKI in advanced NSCLC
patients with positive EGFR T790M: results of a single-arm phase Ib trial,
J. Thorac. Oncol. 16 (2021) S1119–S1120.
[29] C. Zhou, A. Xiong, J. Zhao, W. Li, M. Bi, J. Chen, K. Li, L. Miao, Y. Mao, D. Wang,
7MO oritinib (SH-1028) a third-generation EGFR tyrosine kinase inhibitor in
locally advanced or metastatic NSCLC patients with positive EGFR T790M: results
of a single-arm phase II trial, Ann. Oncol. 33 (2022) S31.
[30] L. Zhao, W. Fu, W. Wu, J. Liu, J. Jin, Method for Preparing Tricyclic Compound as
EGFR Kinase Inhibitor, 2019. CN109705118A.
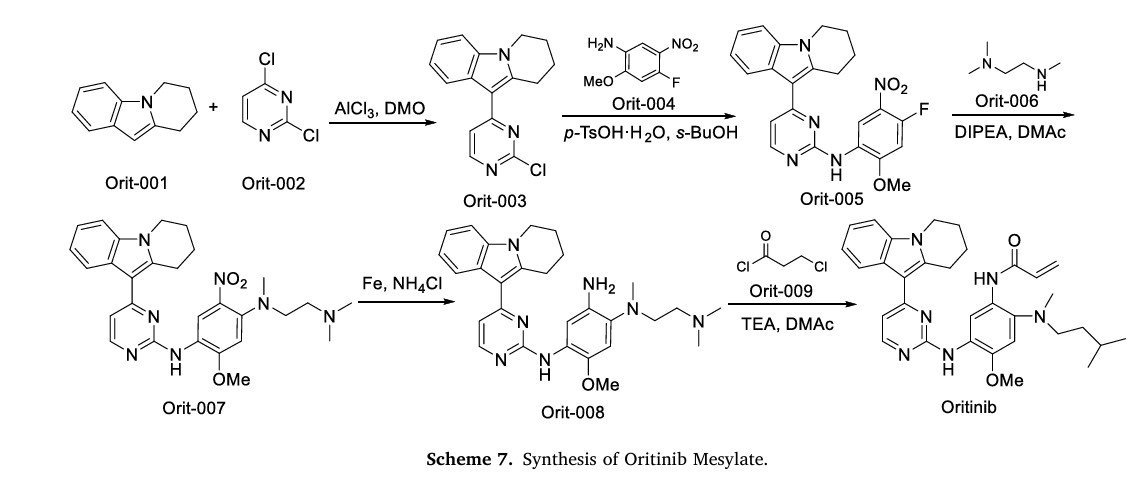
AS ON JUNE2025 4.45 LAKHS VIEWS ON BLOG WORLDREACH AVAILABLEFOR YOUR ADVERTISEMENT
join me on Linkedin
Anthony Melvin Crasto Ph.D – India | LinkedIn
join me on Researchgate
RESEARCHGATE
join me on Facebook
Anthony Melvin Crasto Dr. | Facebook
join me on twitter
Anthony Melvin Crasto Dr. | twitter
+919321316780 call whatsaapp
EMAIL. amcrasto@gmail.com
……
References
- Xiong A, Ren S, Liu H, Miao L, Wang L, Chen J, et al. (October 2022). “Efficacy and Safety of SH-1028 in Patients With EGFR T790M-Positive NSCLC: A Multicenter, Single-Arm, Open-Label, Phase 2 Trial”. Journal of Thoracic Oncology. 17 (10): 1216–1226. doi:10.1016/j.jtho.2022.06.013. PMID 35798241.
- “Rilertinib – Nanjing Sanhome Pharmaceutical”. AdisInsight. Springer Nature Switzerland AG.
| Clinical data | |
|---|---|
| Other names | SH-1028 |
| Identifiers | |
| IUPAC name | |
| CAS Number | 2035089-28-0 |
| PubChem CID | 122666966 |
| ChemSpider | 115007246 |
| UNII | SK593H37SC |
| Chemical and physical data | |
| Formula | C31H37N7O2 |
| Molar mass | 539.684 g·mol−1 |
| 3D model (JSmol) | Interactive image |
| SMILES | |
| InChI | |
- New drugs approved by the NMPA in 2024: Synthesis and clinical applicationsPublication Name: European Journal of Medicinal ChemistryPublication Date: 2025-07-05PMID: 40262297DOI: 10.1016/j.ejmech.2025.117643
- Safety, efficacy, and pharmacokinetics of SH‐1028 in EGFR T790M‐positive advanced non–small cell lung cancer patients: A dose‐escalation phase 1 studyPublication Name: CancerPublication Date: 2023-02-22PMID: 36813747DOI: 10.1002/cncr.34697
- SH-1028, An Irreversible Third-Generation EGFR TKI, Overcomes T790M-Mediated Resistance in Non-Small Cell Lung CancerPublication Name: Frontiers in PharmacologyPublication Date: 2021-04-27PMCID: PMC8111447PMID: 33986687DOI: 10.3389/fphar.2021.665253
- [1]. Luwei Han, et al. SH-1028, An Irreversible Third-Generation EGFR TKI, Overcomes T790M-Mediated Resistance in Non-Small Cell Lung Cancer. Front Pharmacol. 2021 Apr 27;12:665253. [Content Brief]
/////////Oritinib, CHINA 2024, APPROVALS 2024, 2035089-28-0, SH 1028, SK593H37SC, rilertinib, Oritinib mesylate, Nanjing Sanhome Pharmaceutical,
Befotertinib
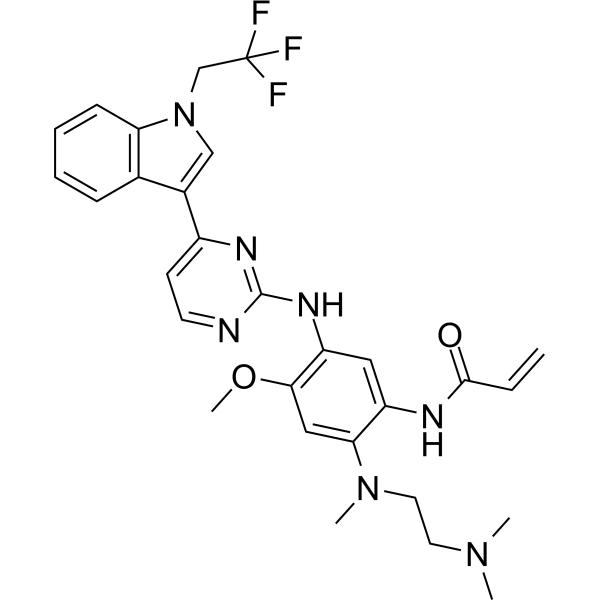
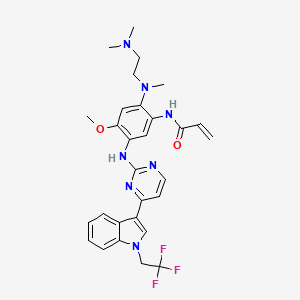
Befotertinib
D-0316, 0XT2CPR891
CAS No. : 1835667-63-4, MESYLATE CAS No. 2226167-02-6
- 2-propenamide, n-(2-((2-(dimethylamino)ethyl)methylamino)-4-methoxy-5-((4-(1-(2,2,2-trifluoroethyl)-1h-indol-3-yl)-2-pyrimidinyl)amino)phenyl)-
- N-(2-((2-(dimethylamino)ethyl)(methyl)amino)-4-methoxy-5-((4-(1-(2,2,2-trifluoroethyl)-1h-indol-3-yl)pyrimidin-2-yl)amino)phenyl)prop-2-enamide
- N-[2-[2-(dimethylamino)ethyl-methylamino]-4-methoxy-5-[[4-[1-(2,2,2-trifluoroethyl)indol-3-yl]pyrimidin-2-yl]amino]phenyl]prop-2-enamide
| Molecular Weight | 567.61 |
|---|---|
| Formula | C29H32F3N7O2 |
Befotertinib (D-0316) is an orally active EGFR tyrosine kinase inhibitor. Befotertinib can inhibit the proliferation of tumor cells. Befotertinib can be used in the research of EGFR T790M-positive non-small cell lung cancer (NSCLC).
Befotertinib is an orally available inhibitor of the epidermal growth factor receptor (EGFR) mutant form T790M, with potential antineoplastic activity. Upon administration, befotertinib specifically binds to and inhibits EGFR T790M, a secondarily acquired resistance mutation, which prevents EGFR-mediated signaling and leads to cell death in EGFR T790M-expressing tumor cells. Compared to some other EGFR inhibitors, befotertinib may have therapeutic benefits in tumors with T790M-mediated drug resistance. EGFR, a receptor tyrosine kinase that is mutated in many tumor cell types, plays a key role in tumor cell proliferation and tumor vascularization.
PAPER
J. Med. Chem. 2017, 60, 6480−6515.
PATENT
WO 2019218987
https://patentscope.wipo.int/search/en/WO2019218987
[0054]
U.S. Publication No. 2017/0355696 A1 describes a method of preparing Compound 4 and various pharmaceutically acceptable salts thereof. The exemplified synthetic process in U. S. Publication No. 2017/0355696 A1 includes a two-step conversion from the aniline compound, corresponding to Compound 1 of this disclosure, into the bismesylate of Compound 4, which has a low yield.
[0055]
As shown herein, representative methods of preparation of Compound 4, or a pharmaceutically acceptable salt, (or alternatively referred to as synthetic methods) , can provide the desired Compound 4, or a pharmaceutically acceptable salt, in improved yield and high purity and can be adapted for large-scale manufacture.
[0056]
In various embodiments, the present invention provides a novel method of preparing Compound 4, or a pharmaceutically acceptable salt thereof. The method typically includes converting a compound of Formula III, or a salt thereof, into compound 4, typically under an elimination reaction condition:
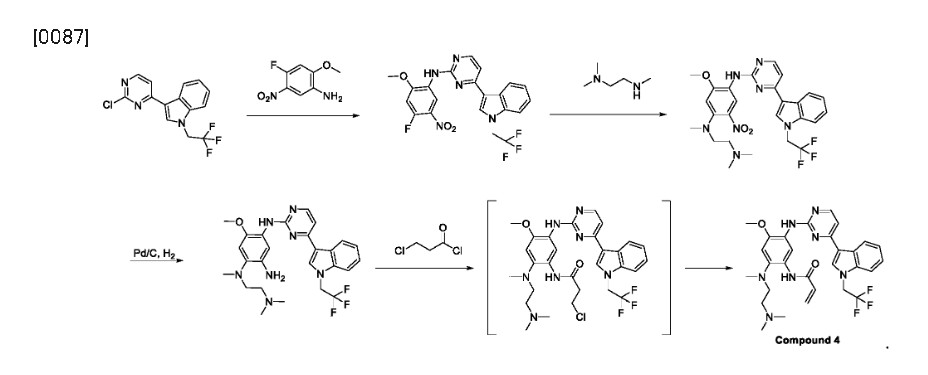
Syn
https://doi.org/10.1021/acs.jmedchem.4c02079
J. Med. Chem. 2025, 68, 2147−2182
Befotertinib (Surmana). Befotertinib (17), an oral, highly selective, third generation epidermal growth factor receptor (EGFR) tyrosine kinase inhibitor (TKI) developed by Betta Pharmaceuticals and InventisBio, was approved in China in May 2023 for the second-line treatment of patients
with locally advanced or metastatic nonsmall cell lung cancer (NSCLC) with positive EGFR T790 M mutation who have disease progression on previous EGFR TKI therapy. 140 139 NSCLC
has a high incidence and disease burden in China, which has spurred the development of multiple EGFR TKIs by Chinese companies.
Achromatography-free process route to befotertinib (17) has been reported in the patent literature by researchers at InventisBio (Scheme 29), although details about scale and yields were not provided.
141 142 The reaction sequence closely follows that of osimertinib, a third generation EGFR inhibitor
that was first approved in 2015 and was covered in our previous review.
Osimertinib and befotertinib share a common backbone, differing only in N-substitution on the indole ring.
Friedel−Crafts arylation of 1H-indole with 2,4-dichloropyrimidine (17.1) gave the 3-pyrimidinyl indole 17.2. The trifluoroethyl moiety in indole 17.4 was introduced via Nalkylation of 17.2 with triflate 17.3. This was followed by an SAr reaction with nitroaniline 17.5 to provide amino pyrimidine 17.6. Next, N,N,N′-trimethylethylenediamine (17.7) displaced the electrophilic aryl fluoride in an SNArreaction to generate intermediate 17.8. The acrylamide moiety was installed using a three-step sequence: hydrogenolytic
reduction of the nitro group to the corresponding aniline, acylation with 3-chloropropanoyl chloride, and immediate elimination to the acrylamide. Mesylate salt formation and crystallization furnished befotertinib mesylate (17) in eight steps from 17.1.
(139) Blair, H. A. Befotertinib: first approval. Drugs 2023, 83, 1433−
1437.
(140) Lau, S. C. M.; Ou, S.-H. I. And still they come over troubled
waters: can Asia’s third-generation EGFR tyrosine kinase inhibitors
(Furmonertinib, Aumolertinib, Rezivertinib, Limertinib, Befotertinib,
SH-1028, and Lazertinib) affect global treatment of EGFR+ NSCLC. J.
Thorac. Oncol. 2022, 17, 1144−1154.
(141) Dai, X.; Jiang, Y. Preparation of pyrimidine derivative and its
pharmaceutical salt as EGFR inhibitors for the treatment of cancer and
other diseases. WO 2019218987, 2019.
(142) Flick, A. C.; Ding, H. X.; Leverett, C. A.; Kyne, R. E.; Liu, K. K.
C.; Fink, S. J.; O’Donnell, C. J. Synthetic approaches to the new drugs
approved during 2015. J. Med. Chem. 2017, 60, 6480−6515.
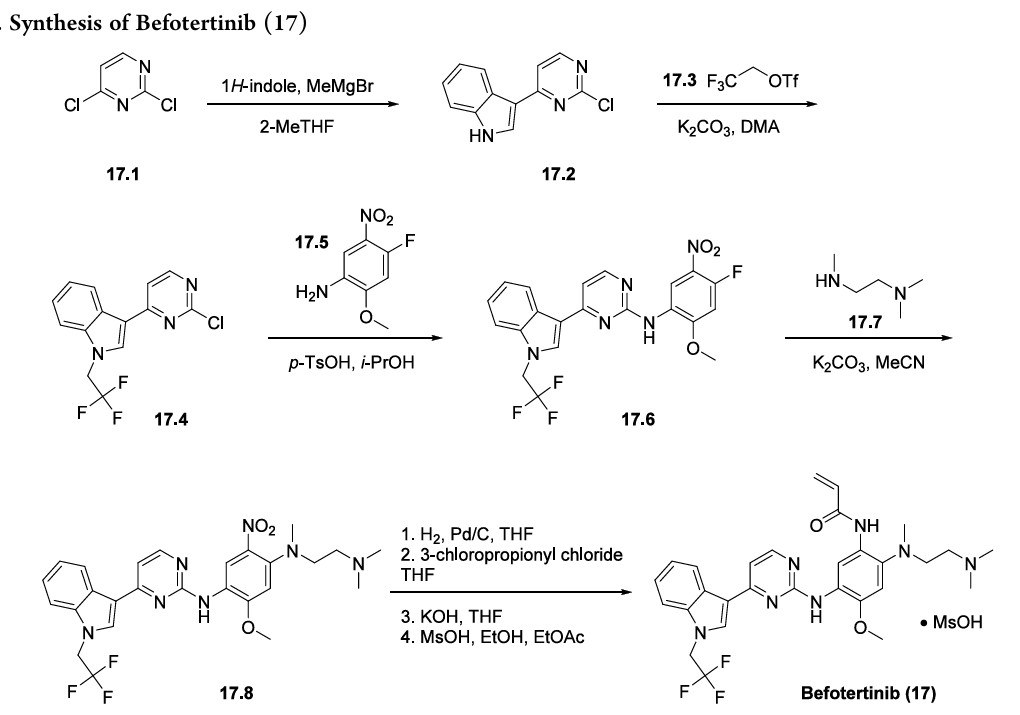
AS ON JUNE2025 4.45 LAKHS VIEWS ON BLOG WORLDREACH AVAILABLEFOR YOUR ADVERTISEMENT
join me on Linkedin
Anthony Melvin Crasto Ph.D – India | LinkedIn
join me on Researchgate
RESEARCHGATE
join me on Facebook
Anthony Melvin Crasto Dr. | Facebook
join me on twitter
Anthony Melvin Crasto Dr. | twitter
+919321316780 call whatsaapp
EMAIL. amcrasto@gmail.com
……
- [1]. Nagasaka M, et, al. Beyond Osimertinib: The Development of Third-Generation EGFR Tyrosine Kinase Inhibitors For Advanced EGFR+ NSCLC. J Thorac Oncol. 2021 May;16(5):740-763. [Content Brief][2]. Blair HA. Befotertinib: First Approval. Drugs. 2023 Oct;83(15):1433-1437. [Content Brief]
/////////Befotertinib, APPROVALS 2023, CHINA 2023, Betta Pharmaceuticals, InventisBio, CANCER, D-0316, D 0316, 0XT2CPR891
Zongertinib
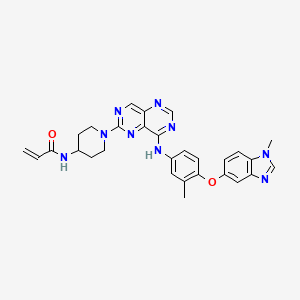
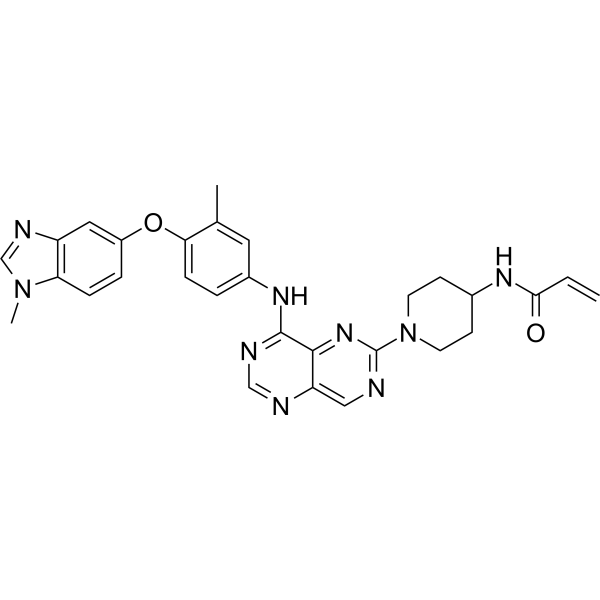
Zongertinib
CAS No. : 2728667-27-2,
BI-1810631, BI1810631
| Molecular Weight | 535.60 |
|---|---|
| Formula | C29H29N9O2 |
FDA 8/8/2025, Hernexeos, To treat adults with unresectable or metastatic non-squamous non-small cell lung cancer whose tumors have HER2 tyrosine kinase domain activating mutations, as detected by an FDA-approved test, and who have received prior systemic therapy
- N-(1-(8-((3-methyl-4-((1-methyl-1H-benzo[d]imidazol-5-yl)oxy)phenyl)amino)pyrimido[5,4-d]pyrimidin-2-yl)piperidin-4-yl)acrylamide
- N-(1-(8-((3-methyl-4-((1-methyl-1H-benzo(d)imidazol-5-yl)oxy)phenyl)amino)pyrimido(5,4-d)pyrimidin-2-yl)piperidin-4-yl)acrylamide
- 884-819-6
Zongertinib is an orally bioavailable inhibitor of the receptor tyrosine kinase human epidermal growth factor receptor 2 (HER2; ErbB2; HER-2), with potential antineoplastic activity. Upon oral administration, zongertinib covalently binds to and inhibits the activity of both wild-type and HER2 mutants, including HER2 mutants with exon 20 insertion (ex20ins) mutations. This prevents HER2-mediated signaling and may lead to cell death in HER2-expressing tumor cells. HER2, a receptor tyrosine kinase overexpressed on a variety of tumor cell types, plays an important role in tumor cell proliferation and tumor vascularization.
REF
Synthesis of zongertinib (N-(1-(8-((3-methyl-4-((1-methyl-1H-benzo[d]imidazol-5-
548 yl)oxy)phenyl)amino)pyrimido[5,4-d]pyrimidin-2-yl)piperidin-4-yl)acrylamide)
Methods
Synthesis of Zongertinib (N-(1-(8-((3-methyl-4-((1-methyl-1H-benzo[d]imidazol-5-yl)oxy)phenyl)amino)pyrimido[5,4-d]pyrimidin-2-yl)piperidin-4-yl)acrylamide)
An overview of the synthetic routes to zongertinib and BI-3999 is shown in Supplementary Fig. S1, and graphical NMR spectra are shown in Supplementary Fig. S2.
3-methyl-4-((1-methyl-1H-benzo[d]imidazol-5-yl)oxy)aniline (500 mg, 1.97 mmol) and 8-chloro-2-(methylthio)pyrimido[5,4-d]pyrimidine hydrochloride (492 mg, 1.97 mmol) were suspended in isopropanol, and the resulting reaction mixture stirred at 50°C for 3 hours, at which time high-performance liquid chromatography–mass spectrometry (HPLC-MS) indicated full conversion. The reaction mixture was concentrated under reduced pressure, and the crude product was redissolved in dichloromethane and washed with aqueous NaHCO3. The organic layer was dried over Na2SO4 and concentrated, and the resulting crude product was purified by column chromatography (SiO2, gradient of 0%–15% methanol in dichloromethane) to afford the product (840 mg).
N-(3-methyl-4-((1-methyl-1H-benzo[d]imidazol-5-yl)oxy)phenyl)-6-(methylthio)pyrimido[5,4-d]pyrimidin-4-amine (860 mg, 90%, 1.80 mmol) was suspended in dichloromethane (30 mL), and the resulting mixture was cooled to 0°C to 5°C. mCPBA (3-chloroperbenzoic acid, 444 mg, 77%, 1.98 mmol) was added portionwise over 1 hour, and the resulting reaction mixture was stirred at room temperature overnight, at which time HPLC-MS indicated full conversion. The reaction mixture was diluted with dichloromethane and washed with aqueous NaHCO3. The organic layer was dried over Na2SO4 and concentrated, and the resulting crude product which was used directly in the next step (767 mg, crude).
N-(3-methyl-4-((1-methyl-1H-benzo[d]imidazol-5-yl)oxy)phenyl)-6-(methylsulfinyl)pyrimido[5,4-d]pyrimidin-4-amine (5.42 g, 80%, 9.73 mmol) was dissolved in N,N-dimethyl formamide (DMF, 50 mL) and diisopropylethylamine (2.8 mL, 16 mmol). 4-Boc-amino-1-piperidine (2.39 g, 11.9 mmol) was added, and the reaction was stirred at 60°C overnight. Then, the reaction mixture was concentrated, and the crude product was used directly in the next step (5.66 g, crude).
Tert-butyl (1-(8-((3-methyl-4-((1-methyl-1H-benzo[d]imidazol-5-yl)oxy)phenyl)amino)pyrimido[5,4-d]pyrimidin-2-yl)piperidin-4-yl)carbamate (5.66 g, 9.73 mmol) was dissolved in dichloromethane (100 mL) and methanol (30 mL). Four mol/L HCl in dioxane (11 mL, 44 mmol) was added, and the resulting reaction mixture was heated to 45°C for 7 hours. HPLC-MS indicated some remaining starting material; therefore, the reaction mixture was stirred at room temperature overnight. Four mol/L HCl in dioxane (1 mL, 0.40 mmol) was added, and the reaction mixture was reheated to 45°C for 4 hours, at which time HPLC-MS indicated full conversion. The reaction mixture was concentrated, and the resulting crude product was purified by column chromatography (SiO2, gradient of 0%–20% methanol in dichloromethane) to afford the product (4.5 g, 70% purity).
1-[8-({3-methyl-4-[(1-methyl-1H-1,3-benzodiazol-5-yl)oxy]phenyl}amino)-[1,3]diazino[5,4-d]pyrimidin-2-yl]piperidin-4-amine (4.5 g, 70%, 6.9 mmol) was suspended in dichloromethane (150 mL) and triethyl amine (4 mL, 28 mmol), and dimethylaminopyridine (115 mg, 0.941 mmol) was added. Then, acroyloyl anhydride (1.36 g, 95%, 10.3 mmol) was added, and the resulting reaction mixture was stirred at room temperature for 1 hour, at which time HPLC-MS indicated full conversion. The reaction mixture was diluted with dichloromethane (50 mL) and washed with aqueous NaHCO3 and brine. The organic layer was dried over Na2SO4 and concentrated, and the resulting crude product was purified by column chromatography (SiO2, gradient of 0%–20% methanol in dichloromethane) to afford the product (2.49 g).
1H NMR (DMSO-d6, 500 MHz) δ 9.58 (s, 1H), 9.08 (s, 1H), 8.39 (s, 1H), 8.19 (s, 1H), 8.10 (d, 1H, J = 7.6 Hz), 7.84 (d, 1H, J = 2.2 Hz), 7.77 (dd, 1H, J = 8.8 Hz, J = 2.2 Hz), 7.57 (d, 1H, J = 8.8 Hz), 7.09 (d, 1H, J = 2.2 Hz), 7.00 (dd, 1H, J = 2.2, 8.5 Hz), 6.89 (d, 1H, J = 8.8 Hz), 6.20 (dd, 1H, J = 10.1, 17.0 Hz), 6.10 (dd, 1H, J = 2.2, 17.0 Hz), 5.6 (dd, 1H, J = 2.2, 9.8 Hz), 4.86 (m, 2H), 3.99 (m, 1H), 3.84 (s, 3H), 3.25 (m, 2H), 2.26 (s, 3H), 1.92 (m, 2H), and 1.43 (m, 2H).
Synthesis of BI-3999 (N-(1-(8-((3-methyl-4-((1-methyl-1H-benzo[d]imidazol-5-yl)oxy)phenyl)amino)pyrimido[5,4-d]pyrimidin-2-yl)piperidin-4-yl)acetamide)
6-(4-aminopiperidin-1-yl)-N-(3-methyl-4-((1-methyl-1H-benzo[d]imidazol-5-yl)oxy)phenyl)pyrimido[5,4-d]pyrimidin-4-amine (100 mg, 208 mmol) and 4-dimethylaminopyridine (2.5 mg, 0.02 mmol) were suspended in 5 mL dichloromethane. Acetic anhydride (25 μL, 0.23 mmol) was added, and the resulting reaction mixture was stirred at room temperature for one hour. Then, the reaction mixture was diluted with dichloromethane and washed with aqueous NaHCO3 and brine. Then, the layers were separated, and the organic layer was dried over MgSO4 and concentrated. The crude product was purified by column chromatography (SiO2, gradient of 0%–10% methanol in dichloromethane) to afford the product (75 mg).
1H NMR (DMSO-d6, 400 MHz) δ 9.58 (s, 1H), 9.07 (s, 1H), 8.39 (s, 1H), 8.17 (s, 1H), 7.88 (d, 1H, J = 7.9 Hz), 7.84 (d, 1H, J = 2.5 Hz), 7.77 (dd, 1H, J = 2.7, 8.7 Hz), 7.57 (d, 1H, J = 8.9 Hz), 7.09 (d, 1H, J = 2.3 Hz), 7.00 (dd, 1H, J = 2.3, 8.6 Hz), 6.89 (d, 1H, J = 8.6 Hz), 4.85 (m, 2H), 3.90 (m, 1H), 3.84 (s, 3H), 3.23 (m, 2H), 2.26 (s, 3H), 1.88 (m, 2H), 1.82 (s, 3H), and 1.38 (m, 2H).
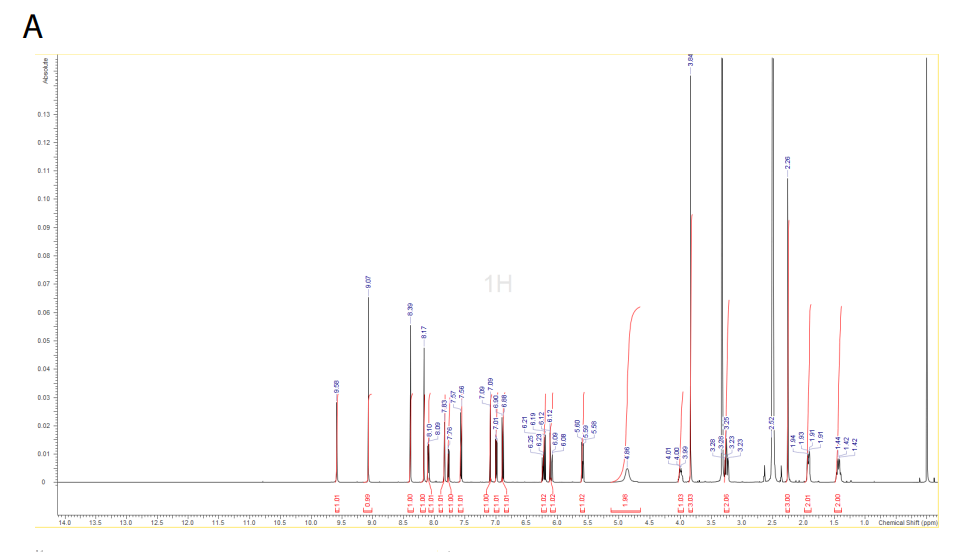
A) 1H NMR spectrum of zongertinib
SYN
https://patentscope.wipo.int/search/en/detail.jsf?docId=WO2021213800&_cid=P10-ME52KD-62836-1
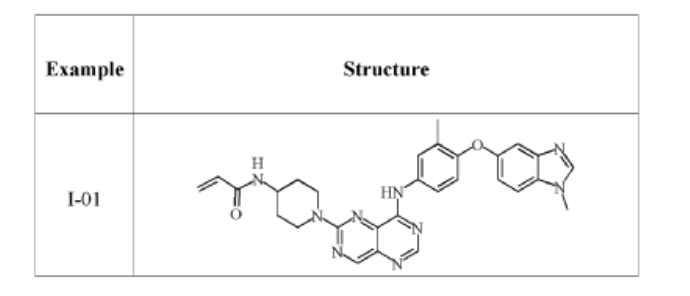
AS ON JUNE2025 4.45 LAKHS VIEWS ON BLOG WORLDREACH AVAILABLEFOR YOUR ADVERTISEMENT
join me on Linkedin
Anthony Melvin Crasto Ph.D – India | LinkedIn
join me on Researchgate
RESEARCHGATE
join me on Facebook
Anthony Melvin Crasto Dr. | Facebook
join me on twitter
Anthony Melvin Crasto Dr. | twitter
+919321316780 call whatsaapp
EMAIL. amcrasto@gmail.com
……
- [1]. WHO Drug Informat ion – World Health Organization (WHO).[2]. Wilding Birgit, et al. Synthesis of diazino-pyrimidines as anticancer agents: World Intellectual Property Organization, WO2021213800. 2021-10-28.[3]. Li S, et al. Emerging Targeted Therapies in Advanced Non-Small-Cell Lung Cancer. Cancers (Basel). 2023 May 24;15(11):2899. [Content Brief]
////////////Zongertinib, Hernexeos, APPROVALS 2025, FDA 2025, lung cancer, BI-1810631, BI1810631, DRH7R67UVL
Olverembatinib

Olverembatinib
1257628-77-5- 3-((1H-pyrazolo[3,4-b]pyridin-5-yl)ethynyl)-4-methyl-N-(4-((4-methylpiperazin-1-yl)methyl)-3-(trifluoromethyl)phenyl)benzamide
- HQP1351
- 4-methyl-N-[4-[(4-methylpiperazin-1-yl)methyl]-3-(trifluoromethyl)phenyl]-3-[2-(1H-pyrazolo[3,4-b]pyridin-5-yl)ethynyl]benzamide
- HQP1351 is under investigation in clinical trial NCT03883100 (A Pivotal Study of HQP1351 in Patients of Chronic Myeloid Leukemia in Accelerated Phase With T315I Mutation).
- 4-methyl-N-[4-[(4-methylpiperazin-1-yl)methyl]-3-(trifluoromethyl)phenyl]-3-[2-(1H-pyrazolo[3,4-b]pyridin-5-yl)ethynyl]benzamide
- D-824
- GZD824
WeightAverage: 532.571
Monoisotopic: 532.219844002, Chemical FormulaC29H27F3N6O
| Molecular Weight | 724.77 |
|---|---|
| Formula | C31H35F3N6O7S2 |
Olverembatinib (GZD824) dimesylate is a potent and orally active pan-Bcr-Abl inhibitor. Olverembatinib dimesylate potently inhibits a broad spectrum of Bcr-Abl mutants. Olverembatinib dimesylate strongly inhibits native Bcr-Abl and Bcr-AblT315I with IC50s of 0.34 nM and 0.68 nM, respectively. Olverembatinib dimesylate has antitumor activity. Olverembatinib (dimesylate) is a click chemistry reagent, it contains an Alkyne group and can undergo copper-catalyzed azide-alkyne cycloaddition (CuAAc) with molecules containing Azide groups.
Olverembatinib is a BCR-ABL1 tyrosine kinase inhibitor developed by Ascentage Pharma. In 2021, it was approved in China “for the treatment of adult patients with TKI-resistant chronic-phase CML (CML-CP) or accelerated-phase CML (CML-AP) harbouring the T315I mutation”.[1][2][3]
SYN
Ren, Xiaomei;Pan, Xiaofen;Zhang, Zhang;Wang, Deping;Lu, Xiaoyun;Li, Yupeng;Wen, Donghai;Long, Huoyou;Luo, Jinfeng;Feng, Yubing;Zhuang, Xiaoxi;Zhang, Fengxiang;Liu, Jianqi;Leng, Fang;Lang, Xingfen;Bai, Yang;She, Miaoqin;Tu, Zhengchao;Pan, Jingxuan;Ding, Ke [Journal of Medicinal Chemistry,2013,vol. 56,# 3,p. 879 – 894]
https://pubs.acs.org/doi/10.1021/jm301581y
PATENT
CN 114163434
https://patentscope.wipo.int/search/en/detail.jsf?docId=CN355399053&_cid=P10-MDPKRT-75688-1
| Example |
| The following examples further illustrate but do not limit the present invention. It should be noted that those skilled in the art can make various modifications and improvements without departing from the inventive concept of the present invention, all of which are included in the scope of protection of the present invention. |
| The specific conditions not disclosed in the experimental methods of the following examples can be selected according to conventional methods and conditions, or according to the product instructions. |
| Unless otherwise specified, “room temperature” in the following examples refers to 20°C to 25°C. The term “h” used herein refers to hours. |
| Example 1 |
| Step 1: |
| |
| Under nitrogen, N-methylpyrrolidone (137.6 g) was heated to 30-35°C to obtain the compound of Formula 1 (14.4 g, 1.3 eq) and the compound of Formula 2 (19.14 g, 1 eq). Bis(triphenylphosphate)palladium dichloride (0.46 g, 0.01 eq) and cuprous iodide (0.113 g, 0.01 eq) were added sequentially. Triethylamine (9.45 g, 1.5 eq) was then added under nitrogen. The reaction mixture was heated to 65-75°C and maintained at this temperature for 2 hours. The reaction process was monitored by liquid chromatography-mass spectrometry. The reaction was terminated when the content of the compound of Formula 2 was ≤0.1%. After completion of the reaction, the reaction solution was cooled to 35-45°C and N-acetyl-L-cysteine (1 g, 0.1 eq) was added directly. The reaction was stirred for 4-5 hours. The resulting product was cooled to room temperature, precipitated with water, centrifuged, and washed with pure water to obtain a crude filter cake. The crude filter cake was vacuum-dried and then slurried with a mixture of ethyl acetate and n-heptane (5 mL of the mixed solvent, wherein the volume ratio of ethyl acetate to n-heptane was 1:1) at a rate of 5 mL per gram of crude filter cake. The resulting slurry was vacuum-dried to yield the compound of Formula 3 with a yield of 85.97% and a purity of 98.2%. |
| The NMR data for the compound of Formula 3 are as follows : 1 H NMR (400 MHz, d-DMSO): δ ppm: 8.93 (1H, d, J = 2.0 Hz); 8.63 (1H, d, J = 2.0 Hz); 8.49 (1H, s); 8.11 (1H, d, J = 2.0 Hz); 7.92 (1H, dd, J = 1.6 Hz; J = 8.0 Hz); 7.52 (1H, d, J = 8.0 Hz); 3.88 (3H, s); 2.59 (3H, s); 1.65 (9H, s). |
| Step 2: |
| |
| Under nitrogen, methanol (160 g) and water (50 g) were sequentially added to the compound of formula 3 (20 g, 1.0 eq). The reaction system was stirred at reflux for 18 hours with process control. The resulting product was cooled to room temperature and filtered to obtain a filter cake (no drying required). Recrystallization was performed by adding 10 times the mass of the filter cake in methanol. The resulting mixture was stirred at 60-70°C for 8-10 hours, then cooled to 40-50°C and subjected to a gradient cooling process at a cooling rate of 5°C per 1 to 1.5 hours to slowly form a solid precipitate. The resulting mixture was filtered, the filter cake was washed with methanol, and vacuum dried to obtain the compound of formula 4 in a 91% yield and 99.7% purity. |
| The NMR data for the compound of Formula 4 are as follows : 1 H NMR (400 MHz, d-DMSO): δ ppm: 8.73 (1H, d, J = 2.0 Hz); 8.52 (1H, t, J = 2.0 Hz); 8.21 (1H, d, J = 2.0 Hz); 8.06 (1H, s); 7.86 (1H, dd, J1 = 2.0 Hz; J2 = 8.0 Hz); 7.49 (1H, dd, J1 = 1.6 Hz; J2 = 7.6 Hz); 3.86 (3H, s); 2.56 (3H, s). |
| Step 3: |
| |
| Under nitrogen, THF (448 mL), compound of formula 4 (29.1 g, 1 eq), and compound of formula 5 (24.6 g, 0.9 eq) were added, stirred, and cooled to -65°C to -60°C. At this temperature, potassium tert-butoxide (19 g x 3) was added in batches every 0.5 h. The reaction process was controlled by liquid phase detection. After 2 hours, the reaction temperature was raised to -5 to 0°C. The reaction solution was washed with purified water, stirred for 0.5-1 hour, washed with brine, and separated to obtain an organic phase. N-acetyl-L-cysteine (11.41 g, 0.7 eq) was added to the organic phase, stirred, washed with brine, neutralized, and concentrated under reduced pressure. The resulting filter cake was washed with purified water and made into a slurry. The resulting product was washed again with purified water and dried under vacuum to obtain compound of formula 6 with a yield of 88.2% and a purity of 98.6%. |
| The NMR data for the compound of formula 6 are as follows : 1 H NMR (400 MHz, d-DMSO): δ ppm: 10.53 (1H, s); 8.75 (d, J = 2.0); 8.53 (d, J = 2.4); 8.24 (1H, s); 8.23 (d, J = 2.4); 8.21 (d, J = 1.6); 8.09 (dd, J1 = 1.6; J2 = 8.4); 7.94 (dd, J1 = 2.0; J2 = 8.0); 7.71 (d, J = 8.8); 7.53 (d, J = 8.0); 3.56 (2H, s); 2.59 (3H, s); 2.34-2.35 (8H, m), 2.16 (3H, s). |
| Its carbon spectrum data are 13 C NMR (100 MHz, d-DMSO): δ ppm: 20.38, 45.65, 52.64, 54.67, 57.41, 88.26, 91.86, 111.76, 113.98, 117.19, 122.14, 123.43, 127.35 (q), 124.30 (q), 128.10, 129.89, 130.49, 131.15, 132.02, 132.13, 132.93, 133.66, 138.15, 143.65, 150.55, 164.64. |
PATENT
CN 101885722
https://patentscope.wipo.int/search/en/detail.jsf?docId=CN84081329&_cid=P10-MDPKML-68458-1
| Example 23 |
| 3-((1H-pyrazolo[3,4-b]pyridine-5-substituted)ethynyl)-4-methyl-N-(4-((4-methylpiperazine-1-substituted)methyl)3-(trifluoromethyl)phenyl)benzamide (D824) |
| (3-((1H-pyrazolo[3,4-b]pyridin-5-yl)ethynyl)-4-methyl-N-(4-((4-methylpiperazin-1-yl)m ethyl)-3-(trifluoromethyl)phenyl)benzamide) |

| The synthesis method is the same as in Example 1. |
| 1 HNMR (400MHz, d-DMSO), δ13.92 (s, 1H), 10.55 (s, 1H), 8.72 (d, J=2.0Hz, 1H), 8.52 (d, J=2.0Hz, 1H), 8.17 (m, 3H), 8.10 (d, J=8.0Hz, 1H), 7.92 (dd, J=8.0, 2.0Hz, 1H), 7.70 (d, J=8.8Hz, 1H), 7.53 (d, J=8.0Hz, 1H), 3.80 (s, 2H), 3.10 (brs, 8H), 2.71 (s, 3H), 2.57 (s, 3H). |
| MS(ESI), m/z: 533, (M + +H + ). |
SYN
Olverembatinib(24) wasdeveloped by Ascentage Pharma as anorally available, third-generation
tyrosinekinase inhibitor (TKI) for the treatment of chronic myeloid leukemia (CML), acute myeloid leukemia, acute lymphoblastic leukemia (ALL), and solid tumors.167 It received its first approval inChina inNovember 2021 and was approved for use in adults with TKI-resistant CML chronicphaseandCML-acceleratephaseharboringtheT315I “gatekeeper” mutation.168 The current mainstay of CML
treatmentiscenteredaroundTKIs;however,resistancetoTKItherapy, often through BCR-ABL1 kinase domain point mutations, remains a challenge for early generation therapies.169Olverembatinibretainsitsefficacybyfunctioningasan ATP-bindingsiteinhibitorofwild-typeBCR-ABL1kinaseand broadly relatedmutants including T315I, which otherwise confers resistance against all first and second generation TKIs.168
Thesynthesisofolverembatinibhasbeenreportedinseveral patents,170−172 aswell as a journal article173 that details the divergentapproachtorelatedanalogues. Inarecentpatent,170 the synthesis of olverembatinib began with a Sonogashira coupling of commercially available alkyne 24.1 with
bromopyridine24.2toaffordester24.3in98%yield(Scheme43). Cleavage of the N-Boc group was accomplished by refluxingcarbamate24.3inaMeOHandwatermixturetogive pyrazole24.4 in91%yield. AfinalKOtBumediatedamide formation with aniline 24.5 resulted in the isolation of
olverembatinib(24) in88%yield.
(167) Dhillon, S. Olverembatinib: First approval. Drugs 2022, 82,
469−475.
(168) Braun, T. P.; Eide, C. A.; Druker, B. J. Response and resistance
to BCR-ABL1-targeted therapies. Cancer Cell 2020, 37, 530−542.
(169) Shoukier, M.; Kubiak, M.; Cortes, J. Review of new-generation
tyrosine kinase inhibitors for chronic myeloid leukemia. Curr. Oncol.
Rep. 2021, 23, 91.
(170) Wen, J.; Feng, J.; Wu, T.; Cai, M.; Teng, S. Preparation
method of alkynyl containing compound and its intermediate. China
Patent CN 114163434, 2022.
(171) Guo, M.; Wen, J.; Teng, S.; Wu, T.; Feng, J. Preparation of
(trifluoromethylphenyl)(pyrazolo[3,4-b]pyridinylethynyl)benzamide
derivative. China Patent CN 113292556, 2021.
(172) Ding, K.; Wang, D.; Pei, D.; Zhang, Z.; Shen, M.; Luo, K.;
Feng, Y. Heterocyclic alkynylbenzene derivatives as cancer cell line
inhibitors and their preparation, pharmaceutical compositions and use
in the treatment of cancer. China Patent CN 101885722, 2010.
(173) Ren, X.; Pan, X.; Zhang, Z.; Wang, D.; Lu, X.; Li, Y.; Wen, D.;
Long, H.; Luo, J.; Feng, Y.; et al. Identification of GZD824 as an
orally bioavailable inhibitor that targets phosphorylated and non
phosphorylated breakpoint cluster region−abelson (Bcr-Abl) kinase
and overcomes clinically acquired mutation-induced resistance against
imatinib. J. Med. Chem. 2013, 56, 879−894.
AS ON JUNE2025 4.45 LAKHS VIEWS ON BLOG WORLDREACH AVAILABLEFOR YOUR ADVERTISEMENT
join me on Linkedin
Anthony Melvin Crasto Ph.D – India | LinkedIn
join me on Researchgate
RESEARCHGATE
join me on Facebook
Anthony Melvin Crasto Dr. | Facebook
join me on twitter
Anthony Melvin Crasto Dr. | twitter
+919321316780 call whatsaapp
EMAIL. amcrasto@gmail.com
……
References
- Dhillon, Sohita (March 2022). “Olverembatinib: First Approval”. Drugs. 82 (4): 469–475. doi:10.1007/s40265-022-01680-9. PMID 35195876. S2CID 247027755.
- Jiang, Qian; Li, Zongru; Qin, Yazhen; Li, Weiming; Xu, Na; Liu, Bingcheng; Zhang, Yanli; Meng, Li; Zhu, Huanling; Du, Xin; Chen, Suning; Liang, Yang; Hu, Yu; Liu, Xiaoli; Song, Yongping; Men, Lichuang; Chen, Zi; Niu, Qian; Wang, Hengbang; Lu, Ming; Yang, Dajun; Zhai, Yifan; Huang, Xiaojun (18 August 2022). “Olverembatinib (HQP1351), a well-tolerated and effective tyrosine kinase inhibitor for patients with T315I-mutated chronic myeloid leukemia: results of an open-label, multicenter phase 1/2 trial”. Journal of Hematology & Oncology. 15 (1): 113. doi:10.1186/s13045-022-01334-z. PMC 9389804. PMID 35982483.
- Jiang, Qian; Huang, Xiaojun; Chen, Zi; Niu, Qian; Shi, Dayu; Li, Zongru; Hou, Yue; Hu, Yu; Li, Weiming; Liu, Xiaoli; Xu, Na; Song, Yongping; Zhang, Yanli; Meng, Li; Hong, Zhenya; Liu, Bingcheng; Zeng, Shan; Men, Lichuang; Li, Yan; Chen, Suning; Xue, Mengxing; Zhu, Huanling; Li, He; Du, Xin; Lou, Jin; Zhang, Xiaohan; Liang, Yang; Dai, Yujun; Lu, Ming; Wang, Hengbang; Ji, Jiao; Yue, Changai; Yang, Dajun; Zhai, Yifan (5 November 2020). “Novel BCR-ABL1 Tyrosine Kinase Inhibitor (TKI) HQP1351 (Olverembatinib) Is Efficacious and Well Tolerated in Patients with T315I-Mutated Chronic Myeloid Leukemia (CML): Results of Pivotal (Phase II) Trials”. Blood. 136 (Supplement 1): 50–51. doi:10.1182/blood-2020-142142. S2CID 228875477.
| Clinical data | |
|---|---|
| Other names | GZD-824; GZD824 |
| Legal status | |
| Legal status | Investigational |
| Identifiers | |
| IUPAC name | |
| CAS Number | 1257628-77-5 |
| PubChem CID | 51038269 |
| IUPHAR/BPS | 10630 |
| DrugBank | DB16185 |
| ChemSpider | 29395146 |
| UNII | KV1M7Q3CBP |
| ChEMBL | ChEMBL2316582 |
| CompTox Dashboard (EPA) | DTXSID301352011 |
| Chemical and physical data | |
| Formula | C29H27F3N6O |
| Molar mass | 532.571 g·mol−1 |
| 3D model (JSmol) | Interactive image |
| SMILES | |
| InChI | |
[1]. Ren X, Pan X, Zhang Z, Identification of GZD824 as an orally bioavailable inhibitor that targets phosphorylated and nonphosphorylated breakpoint cluster region-Abelson (Bcr-Abl) kinase and overcomes clinically acquired mutation-induced resistance against imatinib. J Med Chem. 2013 Feb 14;56(3):879-94. [Content Brief]
//////////Olverembatinib, approvals 2021, china 2021, Ascentage Pharma, cancer, HQP1351, HQP 1351, D-824, D 824, KV1M7Q3CBP, GZD824
Sontigidomide
Sontigidomide
CAS 2560577-69-5
| Molecular Weight | 513.47 |
|---|---|
| Formula | C26H22F3N3O5 |
N-[[2-(2,6-Dioxo-3-piperidinyl)-2,3-dihydro-1-oxo-1H-isoindol-5-yl]methyl]-α-oxo-4-[1-(trifluoromethyl)cyclopropyl]benzeneacetamide
enzeneacetamide, N-[[2-(2,6-dioxo-3-piperidinyl)-2,3-dihydro-1-oxo-1H-isoindol-5-yl]methyl]-α-oxo-4-[1-(trifluoromethyl)cyclopropyl]-
FDD2NVW84X, Sontigidomida
Sontigidomide (Compound 5) is an antineoplastic compound. Sontigidomide inhibits MOLM-13 cell proliferation more than 80% at 1 μM (3 days).
SCHEME
COUPLER………….
MAIN……….
PATENTS
WO2023070120 BioTheryX, Inc.
PATENT
US20200369679
https://patentscope.wipo.int/search/en/detail.jsf?docId=US311579044&_cid=P20-MD87Y5-18242-1
Example 5
Compound I-5: N-((2-(2,6-Dioxopiperidin-3-yl)-1-oxoisoindolin-5-yl)methyl)-2-oxo-2-(4-(1-(trifluoromethyl)cyclopropyl)phenyl)acetamide
Compound I-5 was synthesized as shown in Scheme 5.
To a solution of 3-(5-(aminomethyl)-1-oxoisoindolin-2-yl)piperidine-2,6-dione 8 (80 mg, 0.258 mmol) in DCM (4 mL) at 0° C. was added TEA (52.2 mg, 0.516 mmol). After stirring for 2 min, 2-oxo-2-(4-(1-(trifluoromethyl)cyclopropyl)phenyl)acetyl chloride 13 (71.3 mg, 0.258 mmol) was added and the mixture was stirred at RT for 2 h. After concentration, the residue was purified using prep-HPLC eluting with ACN/H 2O (0.1% TFA) from 10% to 95% to afford compound I-5 (16.1 mg) in 12% yield. MS (ESI) m/z: 514.0 [M+H] +; 1H NMR (400 MHz, DMSO-d 6) δ 10.98 (s, 1H), 9.57 (t, J=6.0 Hz, 1H), 8.03-8.01 (m, 2H), 7.74-7.48 (m, 7H), 5.13-5.09 (m, 1H), 4.59-4.57 (m, 2H), 4.49-4.31 (m, 2H), 2.95-2.87 (m, 1H), 2.63-2.58 (m, 1H), 2.45-2.38 (m, 1H), 2.03-1.99 (m, 1H), 1.43-1.40 (m, 2H), 1.24-1.21 (m, 2H).
- Ketoamides for treating malignancyPublication Number: WO-2023070120-A1Priority Date: 2021-10-22
- Protein-targeting compounds and pharmaceutical compositions thereof, and their therapeutic applicationsPublication Number: US-2020369679-A1Priority Date: 2019-05-24
- Compounds targeting proteins and pharmaceutical compositions thereof, and their therapeutic applicationsPublication Number: WO-2020242960-A1Priority Date: 2019-05-24
- Compounds targeting proteins and pharmaceutical compositions thereof, and their therapeutic applicationsPublication Number: AU-2020283744-A1Priority Date: 2019-05-24
- Targeted protein compound, its pharmaceutical composition and therapeutic applicationPublication Number: CN-114502543-APriority Date: 2019-05-24
- Compounds targeting proteins and pharmaceutical compositions thereof, and their therapeutic applicationsPublication Number: EP-3976623-A1Priority Date: 2019-05-24
- Protein targeting compounds, pharmaceutical compositions thereof and therapeutic applications thereofPublication Number: KR-20220023343-APriority Date: 2019-05-24
- Protein-targeting compounds and pharmaceutical compositions thereof, and their therapeutic applicationsPublication Number: US-11345712-B2Priority Date: 2019-05-24Grant Date: 2022-05-31
- Protein-targeting compounds and pharmaceutical compositions thereof, and their therapeutic applicationsPublication Number: US-2022298172-A1Priority Date: 2019-05-24
////////Sontigidomide, FDD2NVW84X, CANCER, Sontigidomida
..
AS ON JUNE2025 4.45 LAKHS VIEWS ON BLOG WORLDREACH AVAILABLEFOR YOUR ADVERTISEMENT
join me on Linkedin
Anthony Melvin Crasto Ph.D – India | LinkedIn
join me on Researchgate
RESEARCHGATE
join me on Facebook
Anthony Melvin Crasto Dr. | Facebook
join me on twitter
Anthony Melvin Crasto Dr. | twitter
+919321316780 call whatsaapp
EMAIL. amcrasto@gmail.com
……


