
.
Home » Uncategorized (Page 79)
Category Archives: Uncategorized
 DRUG APPROVALS BY DR ANTHONY MELVIN CRASTO
.....FOR BLOG HOME CLICK HERE
Flag and hits
ORGANIC SPECTROSCOPY
Read all about Organic Spectroscopy on
ORGANIC SPECTROSCOPY INTERNATIONAL

SUBSCRIBE

 Subscribe in a reader
Enter your email address:
Delivered by FeedBurner
Subscribe to New Drug Approvals by Email


Recent Comments
 | shivkr2 on SPSR Excellence Award 2025 – S… |
 | Bonkasaurus on Infigratinib phosphate |
 | PALOVAROTENE | ORGAN… on Palovarotene |
| Pfizer just purchase… on Temanogrel | |
| Pfizer To Cash In On… on Temanogrel |
ICH Q3D implemented in the European Pharmacopoeia: Revision of Two General Monographs with Regard to Elemental Impurities

 ICH Q3D implemented in the European Pharmacopoeia: Revision of Two General Monographs with Regard to Elemental Impurities
Two general monographs of the European Pharmacopoeia have been revised and published for comment in the newest “Pharmeuropa” edition. Read more about what you will have to consider in future with regard to the control of elemental impurities in pharmaceutical preparations, APIs and excipients.
see
In a press release dated 30 November 2015, the EDQM announced the revision of two general pharmacopoeial monographs: “Substances for pharmaceutical use” (2034) and “Pharmaceutical preparations” (2619). The decision was taken during the 153rd session of the European Pharmacopoeia Commission; the Commission follows its strategy for implementing the ICH Guideline Q3D “Guideline for Elemental Impurities” in the European Pharmacopoeia. A section “Elemental Impurities” has been added to both monographs which emphasizes that the provisions laid down in General Chapter 5.20 of the Pharmacopoeia (identical in wording with…
View original post 272 more words
GDC-0919; NLG-919; RG-6078

MF C18H22N2O
MW: 282.17321
GDC-0919; NLG-919; RG-6078, GDC0919; GDC-0919; GDC 0919; NLG919; NLG 919; NLG-919; RG6078; RG-6078; RG 6078.
1-cyclohexyl-2-(5H-imidazo[5,1-a]isoindol-5-yl)ethanol
CAS No.1402836-58-1
GDC-0919, also known as NLG919 and RG6078, is an orally available inhibitor of indoleamine 2,3-dioxygenase 1 (IDO1), with potential immunomodulating and antineoplastic activities. Upon administration, NLG919 targets and binds to IDO1, a cytosolic enzyme responsible for the oxidation of the essential amino acid tryptophan into kynurenine. By inhibiting IDO1 and decreasing kynurenine in tumor cells, this agent increases tryptophan levels, restores the proliferation and activation of various immune cells, including dendritic cells (DCs), natural killer (NK) cells, T-lymphocytes, and causes a reduction in tumor-associated regulatory T-cells (Tregs). Activation of the immune system, which is suppressed in many cancers, may induce a cytotoxic T-lymphocyte (CTL) response against the IDO1-expressing tumor cells

- Originator Lankenau Institute for Medical Research
- Developer Genentech; NewLink Genetics Corporation
- Class Antineoplastics; Small molecules
- Mechanism of Action Immunomodulators; Indoleamine-pyrrole 2,3-dioxygenase inhibitors
Phase I Solid tumours
| Patent ID | Date | Patent Title |
|---|---|---|
| US2015210769 | 2015-07-30 | ANTIBODY MOLECULES TO PD-1 AND USES THEREOF |
| US2014066625 | 2014-03-06 | Fused Imidazole Derivatives Useful as IDO Inhibitors |
- 27 Sep 2015 Pharmacokinetics results from a phase-I clinical trial in Solid tumours presented at the European Cancer Congress 2015 (ECC-2015)
- 27 Sep 2015 Positive efficacy and safety results from a phase-I clinical trial in Solid tumours presented at the European Cancer Congress 2015 (ECC-2015)
- 31 Jul 2015 Phase-I clinical trials in Solid tumours (Combination therapy, Late-stage disease, Second-line therapy or greater) in USA (PO) (NCT02471846)
PATENT
http://www.google.com/patents/WO2012142237A1?cl=en

PATENT
US-20160002249-A1 / 2016-01-07
Fused Imidazole Derivatives Useful as IDO Inhibitors
13041-cyclohexyl-2-(5H-imidazo[5,1- a]isoindol-5-yl)ethanol79 1H NMR (a mixture of diastereomers) 1.10-1.37 (m, 6H), 1.66-1.80 (m, 5H), 2.05 (m, 2H), 2.15 (m, 1H), 3.72 (m, 1H), 5.36 and 5.46 (two m, 1H), 7.16 (s, 1H), 7.25 (m, 1H), 7.34 (m, 1H), 7.43 (d, 1H, J = 7.6 Hz), 7.54 (d, 1H, J = 7.6 Hz), 7.80 (s, 1H)
| WO2011056652A1 * | Oct 27, 2010 | May 12, 2011 | Newlink Genetics | Imidazole derivatives as ido inhibitors |
| WO2012142237A1 * | Apr 12, 2012 | Oct 18, 2012 | Newlink Geneticks Corporation | Fused imidazole derivatives useful as ido inhibitors |
| WO2014159248A1 | Mar 10, 2014 | Oct 2, 2014 | Newlink Genetics Corporation | Tricyclic compounds as inhibitors of immunosuppression mediated by tryptophan metabolization |
| US8722720 | Oct 27, 2010 | May 13, 2014 | Newlink Genetics Corporation | Imidazole derivatives as IDO inhibitors |
| US9260434 | Oct 14, 2013 | Feb 16, 2016 | Newlink Genetics Corporation | Fused imidazole derivatives useful as IDO inhibitors |
| US20140066625 * | Oct 14, 2013 | Mar 6, 2014 | Newlink Genetics Corporation | Fused Imidazole Derivatives Useful as IDO Inhibitors |
| US20160002249 * | Jul 8, 2015 | Jan 7, 2016 | Newlink Genetics Corporation | Fused Imidazole Derivatives Useful as IDO Inhibitors |
REFERENCES
Nature Reviews Drug Discovery14,373(2015)doi:10.1038/nrd4658
http://www.ncbi.nlm.nih.gov/pubmed/21517759
http://www.roche.com/irp150128-annex.pdf
/////CRD1152, CRD 1152, CRD-1152, Curadev, Research Collaboration, Licensing Agreement, Develop, Cancer Immunotherapeutic, IDO1 and TDO inhibitors
OC(C1CCCCC1)CC(C2=C3C=CC=C2)N4C3=CN=C4
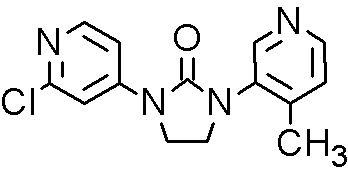
CFG 920, Novartis Scientists team up with Researchers at Aurigene, Bangalore, India,
CFG920,
Inhibitor Of Prostate Cancer With Fewer Cardiac Side Effects
Cas 1260006-20-9
Novartis
Target: CYP17/CYP11B2
Disease: Castration-resistant prostate cancer
MF C14H13ClN4O
MW: 288.0778
Elemental Analysis: C, 58.24; H, 4.54; Cl, 12.28; N, 19.40; O, 5.54
Steroid 17-alpha-hydroxylase inhibitors
CFG920 is a CYP17 inhibitor, is also an orally available inhibitor of the steroid 17-alpha-hydroxylase/C17,20 lyase (CYP17A1 or CYP17), with potential antiandrogen and antineoplastic activities. Upon oral administration, CYP17 inhibitor CFG920 inhibits the enzymatic activity of CYP17A1 in both the testes and adrenal glands, thereby inhibiting androgen production. This may decrease androgen-dependent growth signaling and may inhibit cell proliferation of androgen-dependent tumor cells.
https://clinicaltrials.gov/ct2/show/NCT01647789
NCT01647789: A Study of Oral CFG920 in Patients With Castration Resistant Prostate Cancer2012
- 09 Nov 2015Adverse events, efficacy and pharmacokinetics data from the phase I part of a phase I/II trial in Prostate cancer (Metastatic disease) presented at the 27th AACR-NCI-EORTC International Conference on Molecular Targets and Cancer Therapeutics (AACR-NCI-EORTC-2015)
- 29 Jan 2013Phase-I clinical trials in Prostate cancer in Spain (PO)
- 10 Dec 2012Phase-I clinical trials in Prostate cancer in Canada (PO)
In August 2015, preclinical data were presented at the 250th ACS meeting in Boston, MA. In monkeys, treatment with CFG-920 (3 mg/kg, po) showed good bioavailability with F value of 93%, Tmax of 0.5 h, Cmax of 1382 nM.dn and AUC of 2364 nM.h, while CFG-920 (10 mg/kg, po) showed F value of 183%, Cmax of 1179 nM.dn and Tmax of 1.04 h
Bethany Halford on Twitter: “CFG920 – @Novartis CMOS for …
Bethany Halford on Twitter: “CFG920 – @Novartis CMOS for castration resistant prostate cancer #ACSBoston MEDI 1st disclosures http://t.co/XJJ3tCvpUk”

Novartis is developing CFG-920 (structure shown), an oral CYP17 inhibitor, for the potential treatment of metastatic castration-resistant prostate cancer. In March 2013, a phase I/II trial was initiated and at that time, the study was expected to complete in January 2015; in August 2015, clinical data were presented
2015 250th (August 19) Abs MEDI 341
Discovery of CFG920, a dual CYP17/CYP11B2 inhibitor, for the treatment of castration resistant prostate cancer
American Chemical Society National Meeting and Exposition
Christoph Gaul, Prakash Mistry, Henrik Moebitz, Mark Perrone, Bjoern Gruenenfelder, Nelson Guerreiro, Wolfgang Hackl, Peter Wessels, Estelle Berger, Mark Bock, Saumitra Sengupta, Venkateshwar Rao, Murali Ramachandra, Thomas Antony, Kishore Narayanan, Samiulla Dodheri, Aravind Basavaraju, Shekar Chelur

CHEMISTRY COLLABORATORS
Novartis-Aurigene team: (from left) Brahma Reddy V, Thomas Antony, Murali Ramachandra, Venkateshwar Rao G, Wesley Roy Balasubramanian, Kishore Narayanan, Samiulla DS, Aravind AB, and Shekar Chelur. Not pictured: Björn Grünenfelder, Saumitra Sengupta, Nelson Guerreiro, Andrea Gerken, Mark Perrone, Mark Bock, Wolfgang Hackl, Henrik Möbitz, Peter Wessels, Christoph Gaul, Prakash Mistry, and Estelle Marrer.
Credit: Aurigene

| Aurigene Discovery Technologies Limited, an independent subsidiary of Dr Reddy’s, CSN Murthy is chief executive officer |
Preclinical and clinical studies were performed to evaluate the efficacy of CFG-920, a dual cytochrome P450 (CYP)17 and CYP11B2 dual inhibitor, for the potential treatment of castration resistant prostate cancer. CFG-920 showed potent activity against human CYP17 and CYP11B2 enzymes with IC50 values of 0.023 and 0.034 microM, respectively. In monkeys, treatment with CFG-920 (3 mg/kg, po) showed good bioavailability (93%), Tmax of 0.5 h, Cmax of 1382 nM.dn and AUC of 2364 nM.h, while CFG-920 (10 mg/kg, po) showed F value of 183%, Cmax of 1179 nM.dn and Tmax of 1.04 h. In a phase I, first-in-man study, patients received continuous po dosing of CFG-920 (50 mg, bid) plus prednisone (5 mg) in 28-day cycles. At the time of presentation, CFG-920 was under phase II development.

CFG920
WO 2010149755

Novartis team: (clockwise from left) Wolfgang Hackl, Henrik Möbitz, Peter Wessels, Christoph Gaul, Prakash Mistry, and Estelle Marrer., Credit: Novartis
Prostate cancer is the most commonly occurring cancer in men. Doctors often treat the metastatic stage of the disease by depriving the patient of sex hormones via chemical or surgical castration. But if it progresses far enough, the cancer can survive this therapy, transforming into the castration-resistant form. “Once the cancer becomes castration-resistant, the prognosis is poor,” said Novartis’s Christoph Gaul.
In recent years, CYP17, a bifunctional 17α-hydroxylase/17,20-lyase cytochrome P450 enzyme, has emerged as a target for treating castration-resistant prostate cancer. The enzyme catalyzes the biosynthesis of sex hormones, including testosterone, and blocking it can starve prostate cancer of the androgens it needs to thrive.
Johnson & Johnson’s CYP17 inhibitor, abiraterone acetate (Zytiga), a steroid that binds irreversibly to CYP17, was approved by the Food & Drug Administration in 2011. But Novartis scientists thought they could make a better CYP17 inhibitor, Gaul told C&EN. They teamed up with researchers at Aurigene, in Bangalore, India, and came up with their clinical candidate, CFG920.
Unlike abiraterone, CFG920 isn’t a steroid, and it inhibits CYP17 reversibly. It also reversibly inhibits another cytochrome P450 enzyme, CYP11B2, which is involved in the synthesis of the mineralocorticoids, hormones that regulate cardiac function.
Treating prostate cancer patients by lowering their androgen levels turns out to have negative cardiac side effects: Patients’ lipid metabolism is thrown off and their mineralocorticoid levels jump, leading to increases in blood pressure. Those changes can be stressful for the heart. “If prostate cancer patients don’t die because of the cancer, a lot of times they die because of cardiac disease,” Gaul said.
Because CFG920 also keeps mineralocorticoid levels in check, Novartis is hoping the drug candidate will ameliorate some of the cardiac side effects of inhibiting CYP17. The compound is currently in Phase I clinical trials.
PATENT
WO 2010149755
Example 58
Prύpιn”ation ofI'(2’ChIoroψ}ri(ibi-^’\l)’3’f4’metMψ}τUin’3’yl)-imiJazoliJin’2’θne (5HA)-

Using the same reaction conditions as in Example 14. 1-(4-methyl-pyridin-3-yl)- itnida/olidin-2-onc ().-.!.4b: 600 mg. 3.3898 mmol) uas reacted with 2-chloro-4-iodo- py.idine (974 mg.4.067 mmol). 1 , 4-dioxane (60 mL). copper iodide (65 mg, 0.3398 mmol), /r<w.v-1.2-diamino cycK)hexane (0.12 ml,, 1.0169 mmol) and potassium phosphate (2.15 g, 10.1694 mmol) to afford 810 mg of the product (83% yield).
1H NMR (C1DCI3. 300 Mi l/): 6 8.5-8.4 (m. 211). 8.3 (d. IH), 7.6-7.5 (m, 2H). 7.2 (S. 111). 4.1-3.9 (ni. 4H), 2.35 <s. 3H)
LCVIS puιϊt>: 90.8%. nι-7 – 289.1 (M M)
HPl C: 97.14%
REFERENCES
1: Gomez L, Kovac JR, Lamb DJ. CYP17A1 inhibitors in castration-resistant prostate cancer. Steroids. 2015 Mar;95:80-7. doi: 10.1016/j.steroids.2014.12.021. Epub 2015 Jan 3. Review. PubMed PMID: 25560485; PubMed Central PMCID: PMC4323677.
2: Yin L, Hu Q, Hartmann RW. Recent progress in pharmaceutical therapies for castration-resistant prostate cancer. Int J Mol Sci. 2013 Jul 4;14(7):13958-78. doi: 10.3390/ijms140713958. Review. PubMed PMID: 23880851; PubMed Central PMCID: PMC3742227.
///////CFG-920, CYP17 inhibitor (prostate cancer), Novartis, CFG 920, Novartis scientists, team up , researchers , Aurigene, Bangalore, India,
Novartis Molecule for functionally liver selective glucokinase activators for the treatment of type 2 diabetes


(R)-3-Cyclopentyl-N-(5-methoxy-thiazolo[5,4-b]pyridin-2-yl)-2-[4-(4-methyl-piperazine-1-sulfonyl)-phenyl]-propionamide
3-Cyclopentyl-N-(5-methoxy-thiazolo[5,4-b]pyridin-2-yl)-2-[4-(4-methyl-piperazine-1-sulfonyl)-phenyl]-propionamide
(3-Cyclopentyl-N-(5-methoxy-thiazolo[5,4-b]pyridin-2-yl)-2-[4-(4-methyl-piperazine-1-sulfonyl)-phenyl]-propionamide)
cas 866772-52-3
NVP-LBX192
LBX-192

54 Discovery and Evaluation of NVP-LBX192, a Liver Targeted Glucokinase Activator
Thursday, October 8, 2009: 10:30 AM
Nathan Hale North (Hilton Third Floor)
Glucokinase (GK) activators are currently under investigation by a number of pharmaceutical companies with only a few reaching clinical evaluation. A GK activator has the promise of potentially affecting both the beta-cell of the pancreas, by improving glucose sensitive insulin secretion, as well as the liver, by reducing uncontrolled glucose output and restoring post prandial glucose uptake and storage as glycogen. We will describe our efforts to generate liver selective GK activators which culminated in the discovery of NVP-LBX192 (3-Cyclopentyl-N-(5-methoxy-thiazolo[5,4-b]pyridin-2-yl)-2-[4-(4-methyl-piperazine-1-sulfonyl)-phenyl]-propionamide). This compound activated the GK enzyme in vitro at low nM concentrations and significantly reduced glucose levels during an oral glucose tolerance test in normal as well as diabetic mice.
https://acs.confex.com/acs/nerm09/webprogram/Paper75087.html
| Molecular Formula: | C26H33N5O4S2 |
|---|---|
| Molecular Weight: | 543.70132 g/mol |
Sulfonamide-Thiazolpyridine Derivatives, Glucokinase Activators, Treatment Of Type 2 Diabetes
2009 52 (19) 6142 – 6152
Investigation of functionally liver selective glucokinase activators for the treatment of type 2 diabetes
Journal of Medicinal Chemistry
Bebernitz GR, Beaulieu V, Dale BA, Deacon R, Duttaroy A, Gao JP, Grondine MS, Gupta RC, Kakmak M, Kavana M, Kirman LC, Liang JS, Maniara WM, Munshi S, Nadkarni SS, Schuster HF, Stams T, Denny IS, Taslimi PM, Vash B, Caplan SL
2010 240th (August 22) Medi-198
Glucokinase activators with improved physicochemicalproperties and off target effects
American Chemical Society National Meeting and Exposition
Kirman LC, Schuster HF, Grondine MS et al
2010 240th (August 22) Medi-197
Investigation of functionally liver selective glucokinase activators
American Chemical Society National Meeting and Exposition
Schuster HF, Kirman LC, Bebernitz GC et al
PATENT
http://www.google.com/patents/US7750020
EXAMPLE 1 3-Cyclopentyl-N-(5-methoxy-thiazolo[5,4-b]pyridin-2-yl)-2-[4-(4-methyl-piperazine-1-sulfonyl)-phenyl]-propionamide

A. Phenylacetic Acid Ethyl Ester
A solution of phenylacetic acid (50 g, 0.36 mol) in ethanol (150 mL) is treated with catalytic amount of sulfuric acid (4 mL). The reaction mixture is refluxed for 4 h. The reaction is then concentrated in vacuo. The residue is dissolved in diethyl ether (300 mL) and washed with saturated aqueous sodium bicarbonate solution (2×50 mL) and water (1×100 mL). The organic layer dried over sodium sulfate filtered and concentrated in vacuo to give phenylacetic acid ethyl ester as a colorless oil: 1H NMR (400 MHz, CDCl3) δ 1.2 (t, J=7.2, 3H), 3.6 (s, 2H), 4.1 (q, J=7.2, 2H), 7.3 (m, 5H); MS 165 [M+1]+.
B. (4-Chlorosulfonyl-phenyl)-acetic acid ethyl ester
To a cooled chlorosulfonic acid (83.83 g, 48 mL, 0.71 mol) under nitrogen is added the title A compound, phenylacetic acid ethyl ester (59 g, 0.35 mol) over a period of 1 h. Reaction temperature is brought to RT (28° C.), then heated to 70° C., maintaining it at this temperature for 1 h while stirring. Reaction is cooled to RT and poured over saturated aqueous sodium chloride solution (200 mL) followed by extraction with DCM (2×200 mL). The organic layer is washed with water (5×100 mL), followed by saturated aqueous sodium chloride solution (1×150 mL). The organic layer dried over sodium sulfate, filtered and concentrated in vacuo to give crude (4-chlorosulfonyl-phenyl)acetic acid ethyl ester. Further column chromatography over silica gel (60-120 mesh), using 100% hexane afforded pure (4-chlorosulfonyl-phenyl)-acetic acid ethyl ester as a colorless oil.
C. [4-(4-Methyl-piperazine-1-sulfonyl)-phenyl]-acetic acid ethyl ester
A solution of N-methylpiperazine (9.23 g, 10.21 ml, 0.092 mol), DIEA (13 g, 17.4 mL, 0.10 mol) and DCM 80 mL is cooled to 0° C., and to this is added a solution of the title B compound, (4-chlorosulfonyl-phenyl)-acetic acid ethyl ester (22 g, 0.083 mol) in 50 mL of DCM within 30 min. Reaction mixture stirred at 0° C. for 2 h, and the reaction mixture is washed with water (100 mL), followed by 0.1 N aqueous hydrochloric acid solution (1×200 mL). The organic layer dried over sodium sulfate, filtered and concentrated under vacuo to give crude [4-(4-methyl-piperazine-1-sulfonyl)-phenyl]-acetic acid ethyl ester. Column chromatography over silicagel (60-120 mesh), using ethyl acetate afforded pure [4-(4-methyl-piperazine-1-sulfonyl)-phenyl]-acetic acid ethyl ester as white crystalline solid: 1H NMR (400 MHz, CDCl3) δ 1.3 (t, J=7.4, 3H), 2.3 (s, 3H), 2.5 (m, 4H), 3.0 (br s, 4H), 3.7 (s, 2H), 4.2 (q, J=7.4, 2H), 7.4 (d, J=8.3, 2H), 7.7 (d, J=7.3, 2H); MS 327 [M+1]+.
D. 3-Cyclopentyl-2-[4-(4-methyl-piperazine-1-sulfonyl)-phenyl]-propionic acid ethyl ester
A solution of the title C compound, [4-(4-methyl-piperazine-1-sulfonyl)-phenyl]-acetic acid ethyl ester (15 g, 0.046 mol) in a mixture of THF (60 mL) and DMTP (10 mL) is cooled to −78° C. under nitrogen. The resulting solution is stirred at −78° C. for 45 min and to this is added LDA (25.6 mL, 6.40 g, 0.059 mol, 25% solution in THF/Hexane). A solution of iodomethylcyclopentane (11.60 g, 0.055 mol) in a mixture of DMTP (12 mL) and THF (20 mL) is added over a period of 15 min at −78° C. and reaction mixture stirred at −78° C. for 3 h further, followed by stirring at 25° C. for 12 h. The reaction mixture is then quenched by the dropwise addition of saturated aqueous ammonium chloride solution (50 mL) and is concentrated in vacuo. The residue is diluted with water (50 mL) and extracted with ethyl acetate (3×100 mL). The organic solution is washed with a saturated aqueous sodium chloride (2×150 mL), dried over sodium sulfate, filtered and concentrated in vacuo. Column chromatography over silica gel (60-120 mesh), using 50% ethyl acetate in hexane as an eluent to afford 3-cyclopentyl-2-[4-(4-methyl-piperazine-1-sulfonyl)-phenyl]-propionic acid ethyl ester as a white solid: 1H NMR (400 MHz, CDCl3) δ 0.9-2.1 (m, 11H), 1.2 (t, J=7.1, 3H), 2.3 (s, 3H), 2.5 (br s, 4H), 3.0 (br s, 4H), 3.6 (m, 1H), 4.1 (q, J=7.1, 2H), 7.5 (d, J=8.3, 2H), 7.7 (d, J=8.3, 2H); MS 409 [M+1]+.
E. 3-Cyclopentyl-2-[4-(4-methyl-piperazine-1-sulfonyl)-phenyl]-propionic acid
A solution of the title D compound, 3-cyclopentyl-2-[4-(4-methyl-piperazine-1-sulfonyl)-phenyl]-propionic acid ethyl ester (14 g, 0.034 mol) in methanol:water (30 mL:10 mL) and sodium hydroxide (4.11 g, 0.10 mol) is stirred at 60° C. for 8 h in an oil bath. The methanol is then removed in vacuo at 45-50° C. The residue is diluted with water (25 mL) and extracted with ether (1×40 mL). The aqueous layer is acidified to pH 5 with 3 N aqueous hydrochloric acid solution. The precipitated solid is collected by vacuum filtration, washed with water (20 mL), followed by isopropyl alcohol (20 mL). Finally, solid cake is washed with 100 mL of hexane and dried under vacuum at 40° C. for 6 h to give 3-cyclopentyl-2-[4-(4-methyl-piperazine-1-sulfonyl)-phenyl]-propionic acid as a white solid: 1H NMR (400 MHz, CDCl3) δ 1.1-2.0 (m, 11H), 2.4 (s, 3H), 2.7 (br s, 4H), 3.1 (br s, 4H), 3.6 (m, 1H), 7.5 (d, J=8.3, 2H), 7.6 (d, J=8.3, 2H); MS 381 [M+l]+.
F. 5-Methoxy-thiazolo[5,4-b]pyridin-2-ylamine
A solution of 6-methoxy-pyridin-3-ylamine (5.0 g, 0.0403 mol) in 10 mL of acetic acid is added slowly to a solution of potassium thiocyanate (20 g, 0.205 mol) in 100 mL of acetic acid at 0° C. followed by a solution of bromine (2.5 mL, 0.0488 mol) in 5 mL of acetic acid. The reaction is stirred for 2 h at 0° C. and then allowed to warm to RT. The resulting solid is collected by filtration and washed with acetic acid, then partitioned between ethyl acetate and saturated aqueous sodium bicarbonate. The insoluble material is removed by filtration and the organic layer is evaporated and dried to afford 5-methoxy-thiazolo[5,4-b]pyridin-2-ylamine as a tan solid.
G. 3-Cyclopentyl-N-(5-methoxy-thiazolo[5,4-b]pyridin-2-yl)-2-[4-(4-methyl-piperazine-1-sulfonyl)-phenyl]-propionamide
A solution of the title E compound, 3-cyclopentyl-2-[4-(4-methyl-piperazine-1-sulfonyl)-phenyl]-propionic acid (5 g, 0.013 mol) in DCM (250 mL) is cooled to 0° C. and then charged HOBt hydrate (2.66 g, 0.019 mol), followed by EDCI hydrochloride (6 g, 0.031 mol). The reaction mixture is stirred at 0° C. for 5 h. After that the solution of the title F compound, 5-methoxy-thiazolo[5,4-b]pyridin-2-ylamine (2.36 g, 0.013 mol) and D1EA (8 mL, 0.046 mol) in a mixture of DCM (60 mL) and DMF (20 mL) is added dropwise over 30 min. Reaction temperature is maintained at 0° C. for 3 h, then at RT (28° C.) for 3 days. Reaction is diluted with (60 mL) of water and the organic layer is separated and washed with saturated sodium bicarbonate solution (2×50 mL) followed by water washing (2×50 mL) and saturated sodium chloride aqueous solution (1×150 mL). Finally the organic layer is dried over sodium sulfate, filtered, and evaporated under vacuo. The crude product is purified using column chromatography over silica gel (60-120 mesh), using 40% ethyl acetate in hexane as an eluent to afford 3-cyclopentyl-N-(5-methoxy-thiazolo[5,4-b]pyridin-2-yl)-2-[4-(4-methyl-piperazine-1-sulfonyl)-phenyl]-propionamide as a white solid: 1H NMR (400 MHz, CDCl3) δ 0.9-2.1 (m, 11H), 2.2 (s, 3H), 2.5 (br s, 4H), 3.1 (br s, 4H), 3.7 (m, 1H), 4.0 (s, 3H), 6.8 (d, J=8.8, 1H), 7.5 (d, J=8.3, 2H), 7.7 (d, J=8.3, 2H), 7.8 (d, J=8.8, 1H), 8.6 (s, 1H); MS 617 [M+1]+.
H. 3-Cyclopentyl-N-(5-methoxy-thiazolo[5,4-b]pyridin-2-yl)-2-[4-(4-methyl-piperazine-1-sulfonyl)-phenyl]-propionamide dihydrochloride
The title G compound, 3-cyclopentyl-2-(4-methyl piperazinyl sulfonyl)phenyl-N-(5-methoxy-thiazolo[5,4-b]pyridin-2-yl)propionamide (2.8 g, 0.0051 mol) is added to a cooled solution of 10% hydrochloric acid in isopropanol (3.75 mL). The reaction mixture is stirred at 0° C. for 1 h and then at RT for 2 h. The solid is separated, triturated with 10 mL of isopropanol and collected by vacuum filtration and washed with 50 mL of hexane. The solid is dried at 70° C. for 48 h to afford 3-cyclopentyl-N-(5-methoxy-thiazolo[5,4-b]pyridin-2-yl)-2-[4-(4-methyl-piperazine-1-sulfonyl)-phenyl]-propionamide dihydrochloride as an off white solid.
EXAMPLE 2 (R)-3-Cyclopentyl-N-(5-methoxy-thiazolo[5,4-b]pyridin-2-yl)-2-[4-(4-methyl-piperazine-1-sulfonyl)-phenyl]-propionamide

The title compound is obtained analogously to Example 1 by employing the following additional resolution step:
The racemic title E compound of Example 1,3-cyclopentyl-2-[4-(4-methyl-piperazine-1-sulfonyl)-phenyl]-propionic acid (10 g, 0.026 mol) in 1,4-dioxane (500 mL) is treated in a three necked 1 liter flask, equipped with heating mantle, water condenser, calcium chloride guard tube and mechanical stirrer with 3.18 g (0.026 mol) of (R)-(+)-1-phenylethylamine. This reaction mixture is then refluxed at 100° C. for 1 h. The clear reaction solution is cooled to RT (27° C.) and stirred for 10 h. The crystallized salt is collected by filtration under vacuum, washed with 5 mL of hexane and dried under vacuum to afford salt A.
The salt A is dissolved in 1,4-dioxane (500 mL) and heated at 100° C. for 1 h. The clear reaction solution is cooled to RT (27° C.) and stirred for 10 h. The crystallized product is collected by filtration under vacuum, washed with 50 mL of hexane, and dried under vacuum to afford salt B.
The salt B is dissolved in 1,4-dioxane (290 mL) and heated at 100° C. for 1 h. The clear reaction solution is cooled to RT (27° C.) and stirred for 10 h. The crystallized product is collected by filtration under vacuum, washed with 30 mL of hexane, and dried under vacuum to afford salt C.
The salt C is dissolved in 1,4-dioxane (100 mL) and heated at 100° C. for 1 h. The clear reaction solution is cooled to RT (27° C.) and stirred for 10 h. The crystallized product is collected by filtration under vacuum, washed with 30 ml of hexane, and dried under vacuum to afford salt D.
The salt D is treated with aqueous hydrochloric acid solution (20 mL, 1 mL of concentrated hydrochloric acid diluted with 100 mL of water) and stirred for 5 min. The white solid precipitates out and is collected by vacuum filtration, washed with 10 mL of cold water, 5 mL of isopropanol and 20 mL of hexane, and dried under vacuum to yield the hydrochloride salt of (R)-(−)-3-cyclopentyl-2-[4-(4-methyl-piperazine-1-sulfonyl)-phenyl]-propionic acid, salt E.
The salt E is neutralized by stirring with aqueous sodium bicarbonate solution (10 mL, 1 g of sodium bicarbonate dissolved in 120 mL of water) for 5 min. The precipitated solid is collected by filtration, washed with 10 mL of cold water, 100 mL of hexane, and dried to afford (R)-(−)-3-cyclopentyl-2-[4-(4-methyl-piperazine-1-sulfonyl)-phenyl]-propionic acid: m.p. 202.2-203.4° C.
Alternatively, the title compound may be obtained by the resolution of the racemic title compound of Example 1 using the following preparative chiral HPLC method:
- Column: Chiralcel OD-R (250×20 mm) Diacel make, Japan;
- Solvent A: water:methanol:acetonitrile (10:80:10 v/v/v);
- Solvent B: water:methanol:acetonitrile (05:90:05 v/v/v);
- Using gradient elution: gradient program (time, min/% B): 0/0, 20/0, 50/100, 55/0, 70/0;
- Flow rate: 6.0 mL/min; and
- Detection: by UV at 305 nm.
EXAMPLE 3 (S)-3-Cyclopentyl-N-(5-methoxy-thiazolo[5,4-b]pyridin-2-yl)-2-[4-(4-methyl-piperazine-1-sulfonyl)-phenyl]-propionamide

The title compound is prepared analogously to Example 2.
J MED CHEM 2009, 52, 6142-52
Investigation of Functionally Liver Selective Glucokinase Activators for the Treatment of Type 2 Diabetes
† Novartis Institutes for BioMedical Research, Inc., 100 Technology Square, Cambridge, Massachusetts 02139
‡ Torrent Research Centre, Village Bhat, Gujarat, India
J. Med. Chem., 2009, 52 (19), pp 6142–6152
DOI: 10.1021/jm900839k
http://pubs.acs.org/doi/abs/10.1021/jm900839k

Type 2 diabetes is a polygenic disease which afflicts nearly 200 million people worldwide and is expected to increase to near epidemic levels over the next 10−15 years. Glucokinase (GK) activators are currently under investigation by a number of pharmaceutical companies with only a few reaching early clinical evaluation. A GK activator has the promise of potentially affecting both the β-cells of the pancreas, by improving glucose sensitive insulin secretion, as well as the liver, by reducing uncontrolled glucose output and restoring post-prandial glucose uptake and storage as glycogen. Herein, we report our efforts on a sulfonamide chemotype with the aim to generate liver selective GK activators which culminated in the discovery of 3-cyclopentyl-N-(5-methoxy-thiazolo[5,4-b]pyridin-2-yl)-2-[4-(4-methyl-piperazine-1-sulfonyl)-phenyl]-propionamide (17c). This compound activated the GK enzyme (αKa = 39 nM) in vitro at low nanomolar concentrations and significantly reduced glucose levels during an oral glucose tolerance test in normal mice.


PATENT
EP-1735322-B1
Example 2(R)-3-Cyclopentyl-N-(5-methoxy-thiazolo[5,4-b]pyridin-2-yl)-2-[4-(4-methyl-piperazine-1-sulfonyl)-phenyl]-propionamide
The title compound is obtained analogously to Example 1 by employing the following additional resolution step:
The racemic title E compound of Example 1, 3-cyclopentyl-2-[4-(4-methyl-piperazine-1-sulfonyl)-phenyl]-propionic acid (10 g, 0.026 mol) in 1,4-dioxane (500 mL) is treated in a three necked 1 liter flask, equipped with heating mantle, water condenser, calcium chloride guard tube and mechanical stirrer with 3.18 g (0.026 mol) of (R)-(+)-1-phenylethylamine. This reaction mixture is then refluxed at 100°C for 1 h. The clear reaction solution is cooled to RT (27°C) and stirred for 10 h. The crystallized salt is collected by filtration under vacuum, washed with 5 mL of hexane and dried under vacuum to afford salt A.
The salt A is dissolved in 1,4-dioxane (500 mL) and heated at 100°C for 1 h. The clear reaction solution is cooled to RT (27°C) and stirred for 10 h. The crystallized product is collected by filtration under vacuum, washed with 50 mL of hexane, and dried under vacuum to afford salt B.
The salt B is dissolved in 1,4-dioxane (290 mL) and heated at 100°C for 1 h. The clear reaction solution is cooled to RT (27°C) and stirred for 10 h. The crystallized product is collected by filtration under vacuum, washed with 30 mL of hexane, and dried under vacuum to afford salt C.
The salt C is dissolved in 1,4-dioxane (100 mL) and heated at 100°C for 1 h. The clear reaction solution is cooled to RT (27°C) and stirred for 10 h. The crystallized product is collected by filtration under vacuum, washed with 30ml of hexane, and dried under vacuum to afford salt D.
The salt D is treated with aqueous hydrochloric acid solution (20 mL, 1 mL of concentrated hydrochloric acid diluted with 100 mL of water) and stirred for 5 min. The white solid precipitates out and is collected by vacuum filtration, washed with 10 mL of cold water, 5 mL of isopropanol and 20 mL of hexane, and dried under vacuum to yield the hydrochloride salt of (R)-(-)-3-cyclopentyl-2-[4-(4-methyl-piperazine-1-sulfonyl)-phenyl]-propionic acid, salt E.
The salt E is neutralized by stirring with aqueous sodium bicarbonate solution (10 mL, 1 g of sodium bicarbonate dissolved in 120 mL of water) for 5 min. The precipitated solid is collected by filtration, washed with 10 mL of cold water, 100 mL of hexane, and dried to afford (R)-(-)-3-cyclopentyl-2-[4-(4-methyl-piperazine-1-sulfonyl)-phenyl]-propionic acid: m.p. 202.2-203.4°C.
Alternatively, the title compound may be obtained by the resolution of the racemic title compound of Example 1 using the following preparative chiral HPLC method:
- Column: Chiralcel OD-R (250 x 20 mm) Diacel make, Japan;
- Solvent A: water:methanol:acetonitrile (10:80:10 v/v/v);
- Solvent B: water:methanol:acetonitrile (05:90:05 v/v/v);
- Using gradient elution: gradient program (time, min / %B): 0/0, 20/0, 50/100, 55/0, 70/0;
- Flow rate: 6.0 mL/min; and
- Detection: by UV at 305 nm.
REFERENCES
US 7750020
WO-2005095418-A1
US-20080103167-A1
| Patent ID | Date | Patent Title |
|---|---|---|
| US2015218151 | 2015-08-06 | NOVEL PHENYLACETAMIDE COMPOUND AND PHARMACEUTICAL CONTAINING SAME |
| US7750020 | 2010-07-06 | Sulfonamide-Thiazolpyridine Derivatives As Glucokinase Activators Useful The Treatment Of Type 2 Diabetes |
///NOVARTIS, DIABETES, Sulfonamide-Thiazolpyridine Derivatives, Glucokinase Activators, Treatment Of Type 2 Diabetes, 866772-52-3, Novartis Molecule, functionally liver selective glucokinase activators, treatment of type 2 diabetes , NVP-LBX192, LBX-192
c1(sc2nc(ccc2n1)OC)NC(C(c3ccc(cc3)S(=O)(=O)N4CCN(CC4)C)CC5CCCC5)=O
RP 6530, Tenalisib


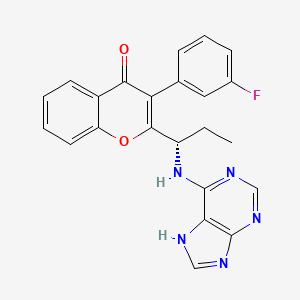
(S)-2-(l-(9H-purin-6-ylamino)propyl)-3-(3-fluorophenyl)-4H-chromen-4-one (Compound A1 is RP 6530).

RP 6530
RP 6530, RP6530, RP-6530
Tenalisib
RP6530-1401, NCI-2015-01804, 124584, NCT02567656
(S)-2-(l-(9H-purin-6-ylamino)propyl)-3-(3-fluorophenyl)-4H-chromen-4-one
3-(3-fluorophenyl)-2-[(1S)-1-(7H-purin-6-ylamino)propyl]chromen-4-one
MW415.4, C23H18FN5O2
CAS 1639417-53-0, 1693773-94-2
A PI3K inhibitor potentially for the treatment of hematologic malignancies.
An inhibitor of phosphoinositide-3 kinase (PI3K) δ/γ isoforms and anti-cellular proliferation agent for treatment of hematol. malignancies
Rhizen Pharmaceuticals is developing RP-6530, a PI3K delta and gamma dual inhibitor, for the potential oral treatment of cancer and inflammation In November 2013, a phase I trial in patients with hematologic malignancies was initiated in Italy ]. In September 2015, a phase I/Ib study was initiated in the US, in patients with relapsed and refractory T-cell lymphoma. At that time, the study was expected to complete in December 2016
PATENTS……..WO 11/055215 , WO 12/151525.
Inventors
| Inventors | Meyyappan Muthuppalaniappan, Srikant Viswanadha, Govindarajulu Babu, Swaroop Kumar V.S. Vakkalanka, |
| Incozen Therapeutics Pvt. Ltd., Rhizen… |
View original post 3,692 more words
RP 6503, Novartis to develop and commercialize Rhizen’s inhaled dual PI3K-delta gamma inhibitor

RP 6503
phase 1

RP 6503
| Molecular Formula: | C30H24F2N6O5S |
|---|---|
| Molecular Weight: | 618.610566 g/mol |
Mass: 619.1 (M++l). MP: 175-178° C Specific optical rotation (C=l in chloroform, at 25°C) : [a]D = + 147.16.

A1
RP 6503
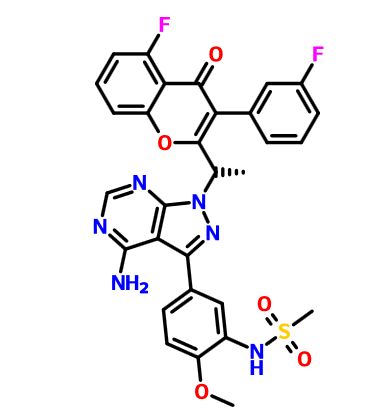
(S)-N-(5-(4-amino-l-(l-(5-fluoro-3-(3-fluorophenyl)-4-oxo-4H-chromen-2-yl) ethyl)-lH-pyrazolo[3,4-d]pyrimidin-3-yl)-2-methoxyphenyl)methanesulfonamide
(S)-N-[5-[4-amino-1-[1-[5-fluoro-3-(3-fluorophenyl)-4-oxochromen-2-yl]ethyl]pyrazolo[3,4-d]pyrimidin-3-yl]-2-methoxyphenyl]methanesulfonamide


Novartis to develop and commercialize Rhizen’s inhaled dual PI3K-delta gamma inhibitor and related compounds worldwide
The immune pipeline includes ‘dual PI3K inhibitors for various indications’ licensed to Novartis
‘inhaled dual inhibitor’,
Phosphoinositide-3 kinase delta inhibitor; Phosphoinositide-3 kinase gamma inhibitor
WO2011055215A2 and WO2012151525A1 and U.S. Publication Nos. US20110118257 and US20120289496
Rhizen Pharmaceuticals Sa INNOVATOR
| Incozen Therapeutics Pvt. Ltd., Rhizen Pharmaceuticals Sa |
PATENT
http://www.google.com/patents/WO2011055215A2?cl=en
PATENT
http://www.google.com/patents/WO2012151525A1?cl=en
scheme 1A:
Ste -1

Step-2

Scheme 2

SCHEME 3

SCHEME4

List of Intermediates


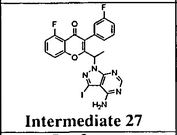
Intermediate 27: 2-( l -(4-amino-3-iodo-lH-pyrazolo[3,4-d]pyrimidin- l – yl)ethyl)-5-fluoro-3-(3-fluorophenyl)-4H-chromen-4-one: To a solution of 3-iodo- l H- pyrazolo[3,4-d]pyrimidin-4-amine (0.800 g, 2.88 mmol) in DMF (5 ml), potassium carbonate (0.398 g, 2.88 mmol) was added and stirred at RT for 30 min. To this mixture intermediate 22 (0.500 g, 1.44 mmol) was added and stirred for 12h. The reaction mixture was diluted with water and extracted with ethyl acetate. The organic layer was dried over sodium sulphate and concentrated under reduced pressure. The crude product was purified by column chromatography with methanol: dichloromethane to afford the title compound as a off-white solid (0.300 g, 38%). Ή-NMR (5 ppm, DMSO-d63, 400 MHz): 8.02 (s, 1 H), 7.94 (s, 1 H), 7.84 (dt, J = 8.4,5.7 Hz, 1H), 7.47 (d, 7 = 8.6 Hz, 1H), 7.29 (m, 3H), 7.09 (dt, 7 = 8.8,2.3 Hz, 1 H), 6.87 (s, 2H), 5.88 (q, 7 = 7.0 Hz, 1H), 1.82 (d, 7 = 7.0 Hz, 3H).
SYNTHESIS
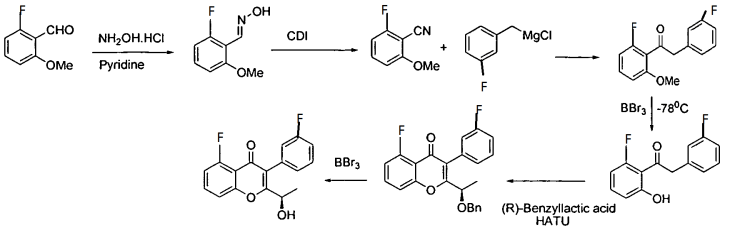
MAIN PART
PATENT
http://www.google.com/patents/WO2015198289A1?cl=en
Prashant Kashinath Bhavar, Swaroop Kumar Venkata Satya VAKKALANKA
The present invention relates to a selective dual delta (δ) and gamma (γ) PI3K protein kinase modulator (S)-N-(5-(4-amino-1-(1-(5-fluoro-3-(3-fluorophenyl)-4-oxo-4H- chromen-2-yl)ethyl)-1H-pyrazolo[3,4-d]pyrimidin-3-yl)-2-methoxyphenyl) methane sulfonamide, methods of preparing them, pharmaceutical compositions containing them and methods of treatment, prevention and/or amelioration of PI3K kinase mediated diseases or disorders with them.
compound of formula (Al):
(Al).
The process comprises the steps of:
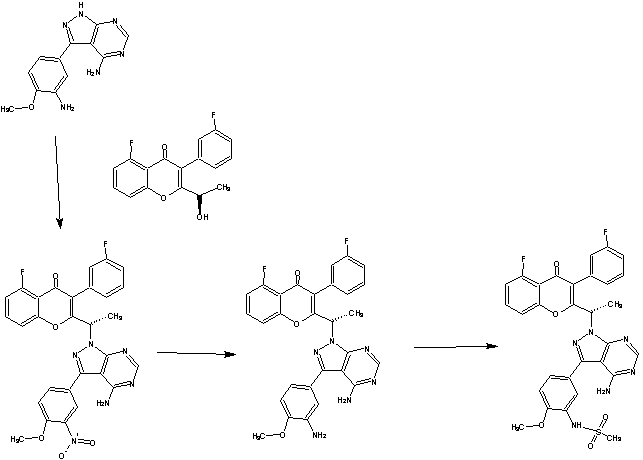
(a) subjecting (R)-5-fluoro-3-(3-fluorophenyl)-2-(l-hydroxyethyl)-4H-chromen-4-one:

to a Mitsunobu reaction with 3-(4-methoxy-3-nitrophenyl)-lH-pyrazolo[3,4-d]pyrimidin-4-amine:

(for example, in the presence of triphenylphosphine and diisopropylazodicarboxylate) to give (S)-2-(l-(4-amino-3-(4-methoxy-3-nitrophenyl)-lH-pyrazolo[3,4-d]pyrimidin-l-yl)ethyl)-5-fluoro-3-(3-fluorophenyl)-4H-chromen-4-one (Intermediate 3):

Intermediate 3;
(b) reducing Intermediate 3, for example with a reducing agent such as Raney Ni, to give (S)-2-(l-(4-amino-3-(3-amino-4-methoxyphenyl)-lH-pyrazolo[3,4-d]pyrimidin- l-yl)ethyl)-5-fluoro-3-( -fluorophenyl)-4H-chromen-4-one (Intermediate 4):

Intermediate 4;
The intermediates described herein may be prepared by the methods described in International Publication Nos. WO 11/055215 and WO 12/151525, both of which are hereby incorporated by reference.
Intermediate 1: N-(5-bromo-2-methoxyphenyl)methanesulfonamide:
To a solution of 5-bromo-2-methoxyaniline(1.00 g, 4.94 mmol) in dichloromethane (10 ml), pyridine (0.800 ml, 9.89 mmol) was added and cooled to 0°C. Methane sulphonyl chloride (0.40 ml, 5.19 mmol) was added and stirred for 30 min. The reaction mixture was quenched with water, extracted with ethyl acetate, dried over anhydrous sodium sulphate and concentrated under reduced pressure. The crude product was chromatographed with ethyl acetate : petroleum ether to afford the title compound as a reddish solid (1.20 g, 87%).
Intermediate 2: N-(2-methoxy-5-(4,4,5,5-tetramethyl-l,3,2-dioxaborolan-2-yl)phenyl)methanesulfonamide: Potassium acetate (0.841 g, 8.57 mmol) and bis(pinacolato)diboron (1.190 g, 4.71 mmol) were added to a solution of intermediate 1 (1.20 g, 4.28 mmol) in dioxane (17.5 ml) and the solution was degassed for 30 min.[l, -Bis(diphenylphosphino)ferrocene]dichloro palladium(II).CH2Ci2 (0.104 g, 0.128 mmol) was added under nitrogen atmosphere and heated to 80°C. After 2h the
reaction mixture was filtered through celite and concentrated. The crude product was purified by column chromatography with ethyl acetate : petroleum ether to afford the title compound as a yellow solid (1.00 g, 71%).JH-NMR (δ ppm, CDCb, 400 MHz): 7. 91 (d, / = 1.2Hz, 1H), 7. 62 (dd, / = 8.1, 1.2Hz, 1H), 6. 92 (d, / = 8.1Hz, 1H), 6.73 (s, 1H), 3.91 (s, 3H), 2.98 (s, 3H), 1.32 (s, 12H).
Intermediate 3: (S)-2-(l-(4-amino-3-(4-methoxy-3-nitrophenyl)-lH-pyrazolo[3,4-d]pyrimidin-l-yl)ethyl)-5-fluoro-3-(3-fluorophenyl)-4H-chromen-4-one: (S)-2-(l-(4-amino-3-(4-methoxy-3-nitrophenyl)-lH-pyrazolo[3,4-d]pyrimidin-l-yl)ethyl)-5-fluoro-3-(3-fluorophenyl)-4H-chromen-4-one: To a solution of (R)-5-fluoro-3-(3-fluorophenyl)-2-(l-hydroxyethyl)-4H-chromen-4-one (0.500 g, 1.64 mmol) in THF (5 ml), 3-(4-methoxy-3-nitrophenyl)-lH-pyrazolo[3,4-d]pyrimidin-4-amine (0.564 g, 1.97 mmol) and triphenylphosphine (0.649 g, 2.47 mmol) were added followed by the addition of diisopropylazodicarboxylate (0.50 ml, 2.47 mmol). ((R)-5-fluoro-3-(3-fluorophenyl)-2-(l-hydroxyethyl)-4H-chromen-4-one can be prepared as described for Intermediates 23, 25, and 26 in International Publication No. WO 2012/0151525.). After 4h at room temperature, the mixture was concentrated and the residue was purified by column chromatography with ethyl acetate : petroleum ether to afford the title compound as a brown solid (0.270 g, 29%). JH-NMR (δ ppm, DMSO-d6, 400 MHz): 8.04 (s, 1H), 7.83 (m, 1H), 7.63-7.50 (m, 3H), 7.29 (m, 2H), 7.06 (dt, J = 8.7,2.2Hz, 1H), 6.94 (m, 2H), 6.75 (dd, J = 8.1,2.1Hz, 1H), 5.95 (q, J = 7.0Hz, 1H), 4.98 (s, 2H), 3.81 (s, 3H), 1.86 (d, J = 7.0 Hz, 3H).
[109] Intermediate 4: (S)-2-(l-(4-amino-3-(3-amino-4-methoxyphenyl)-lH-pyrazolo[3,4-d]pyrimidin-l-yl)ethyl)-5-fluoro-3-(3-fluorophenyl)-4H-chromen-4-one:
(S)-2-(l-(4-amino-3-(3-amino-4-methoxyphenyl)-lH-pyrazolo[3,4-d]pyrimidin-l-yl)ethyl)-5-fluoro-3-(3-fluorophenyl)-4H-chromen-4-one : To a solution of Intermediate 3 (0.260 g, 0.455 mmol) in ethanol (5 ml), Raney Ni (0.130 g) was added and hydrogeneated at 20psi at 50°C for 24h. The reaction mixture was passed through celitepad and concentrated to afford the title compound as a brown solid (0.150 g, 60%). Mass : 540.8 (M+).
Example A
N-(5-(4-amino-l-(l-(5-fluoro-3-(3-fluorophenyl)-4-oxo-4H-chromen-2-yl)ethyl)-lH- pyrazolo[3,4-d]pyrimidin-3-yl)-2-methoxyphenyl)methanesulfonamide
To a solution of 2-(l-(4-amino-3-iodo-lH-pyrazolo[3,4-d]pyrimidin-l-yl)ethyl)-5-fluoro-3-(3-fluorophenyl)-4H-chromen-4-one (0.200 g, 0.366 mmol) in DME (2.1 ml) and water (0.67 ml), intermediate 2 (0.179 g, 0.550 mmol) and sodium carbonate (0.116 g, 1.10 mmol) were added and the system was degassed for 30 min. (2-(l-(4-amino-3-iodo-lH^yrazolo[3,4-d]pyrimidin-l-yl)ethyl)-5-fluoro-3-(3-fluorophenyl)-4H-chromen-4-one can be prepared as described for Intermediates 23, 25, and 26 in International Publication No. WO 2012/0151525). Bis(diphenylphosphino) ferrocene]dichloropalladium(II) (0.059 g, 0.075 mmol) was added and kept under microwave irradiation (microwave power = 100W, temperature = 100 °C) for 45 min. The reaction mixture was Celite filtered, concentrated and extracted with ethyl acetate. The organic layer was dried over sodium sulphate and concentrated under reduced pressure. The crude product was purified by column chromatography with methanol: dichloromethane to afford the title compound as a brown solid (0.080 g, 35%). MP: 216-218 °C. ¾-NMR (δ ppm, CDCb, 400 MHz): 8.20 (s, 1H), 7.73 (s, 1H), 7.53 (m, 2H), 7.31 (m, 2H), 7.07-6.73 (m, 6H), 6.07 (q, / = 6.2 Hz, 1H), 3.98 (s, 3H), 3.14 (s, 3H), 2.01 (d, / = 6.0Hz, 3H).
Example Al and A2
Method A
(S)-N-(5-(4-amino-l-(l-(5-fluoro-3-(3-fluorophenyl)-4-oxo-4H-chromen-2-yl)ethyl)- lH-pyrazolo[3,4-d]pyrimidin-3-yl)-2-methoxyphenyl)methanesulfonamide
and (R)-N-(5-(4-amino-l-(l-(5-fluoro-3-(3-fluorophenyl)-4-oxo-4H-chromen-2- yl)ethyl)-lH-p anesulfonamide

The two enantiomerically pure isomers were separated by preparative SFC (supercritical fluid) conditions from N-(5-(4-amino-l-(l-(5-fluoro-3-(3-fluorophenyl)-4-oxo-4H-chromen-2-yl)ethyl)-lH-pyrazolo[3,4-d]pyrimidin-3-yl)-2-methoxyphenyl)methanesulfonamide (0.500 g) on a CHIRALPAK AS-H column (250 x 30 mm; 5μπι) using methanol : CO2 (55:45) as the mobile phase at a flow rate of 80g / min.
Example Al (S-isomer): Brown solid (0.247 g). Enantiomeric excess: 97.4%. Retention time: 2.14 min. Mass: 619.1 (M++l). MP: 156-158° C.
Example A2 (R-isomer): Brown solid (0.182 g). Enantiomeric excess: 99.3%. Retention t: 3.43 min. Mass: 619.1 (M++l). MP: 168-171° C.
Method Al
(S)-N-(5-(4-amino-l-(l-(5-fluoro-3-(3-fluorophenyl)-4-oxo-4H-chromen-2-yl)ethyl)- lH-pyrazolo[3,4-d]pyrimidin-3-yl)-2-methoxyphenyl)methanesulfonamide
and (R)-N-(5-(4-amino-l-(l-(5-fluoro-3-(3-fluorophenyl)-4-oxo-4H-chromen-2- yl)ethyl)-lH-p anesulfonamide

The two enantiomerically pure isomers were separated by preparative SFC (supercritical fluid) conditions from N-(5-(4-amino-l-(l-(5-fluoro-3-(3-fluorophenyl)-4-oxo-4H-chromen-2-yl)ethyl)-lH-pyrazolo[3,4-d]pyrimidin-3-yl)-2-methoxyphenyl) methanesulfonamide (15.0 g) on a CHIRALPAK AS-H column (250 x 20 mm; 5μπι) using methanol : CO2 (45:55) as the mobile phase at a flow rate of 120g / min.
Example Al (S-isomer): Enantiomeric excess: 100 %. Retention time: 2.21 min. Mass: 619.1 (M++l). MP: 175-178° C Specific optical rotation (C=l in chloroform, at 25°C) : [a]D = + 147.16.
Example A2 (R-isomer): Enantiomeric excess: 99.3%. Retention t: 3.72 min. Mass: 619.1 (M++l). MP: 154-157° C. Specific optical rotation (C=l in chloroform, at 25°C) : [a]D = – 159.54.
Method B
Example Al
(S)-N-(5-(4-amino-l-(l-(5-fluoro-3-(3-fluorophenyl)-4-oxo-4H-chromen-2-yl)ethyl)- lH-pyrazolo[3,4-d]pyrimidin-3-yl)-2-methoxyphenyl)methanesulfonamide
To a solution of Intermediate 4 (0.500 g, 0.923 mmol) in dichloromethane (5 ml) cooled to 0°C, pyridine (0.200 ml, 1.84 mmol) was added and stirred for 10 min. Methanesulphonyl chloride (0.100 ml, 0.923 mmol) was added stirred for 30 min. The reaction mixture was quenched with water, extracted with dichloromethane and dried over sodium sulphate. The crude product was column chromatographed with methanol : dichloromethane to afford the title compound as an off-white solid (0.240 g, 42%). MP: 211-213°C. ¾-NMR (δ ppm, DMSO-d6, 400 MHz): 9.15 (s, 1H), 8.06 (s, 1H), 7.83 (m, 1H), 7.49 (m, 4H), 7.28 (m, 4H), 7.08 (dt, / = 8.6, 1.7 Hz, 1H), 6.92 (s, 2H), 5.98 (q, / = 6.9 Hz, 1H), 3.88 (s, 3H), 2.99 (s, 3H), 1.88 (d, / = 7.0 Hz, 3H). Enantiomeric excess: 85.4% as determined by HPLC on a chiralpak AS-3R column, enriched in the fast eluting isomer (retention time = 7.46 min.).

CLIPS
La Chaux-de-Fonds, Switzerland, Sept. 6, 2013 — La Chaux-de-Fonds, Switzerland (6 September 2013): Rhizen Pharmaceuticals S.A. announces a scientific poster presentation on the pre-clinical characterization of its lead calcium release activated channel (CRAC) inhibitor, RP3128, for the treatment of respiratory disorders and an oral presentation on the pharmacological profile of its novel, dual Phosphoinositide-3 kinase (PI3K) delta/gamma inhibitor, RP6503, in the pulmonary disease systems, at the European Respiratory Society Annual Congress (ERS), to be held from 7-11 September 2013, at Barcelona, Spain.
RP6503 is a novel, potent and selective inhibitor of the delta and gamma isoforms of PI3K. It is to be delivered via the inhalation route and has a long duration of action along with excellent PI3K isoform selectivity, which is expected to result in better safety. RP3128 has been optimized with high potency for CRAC channel inhibition, selectivity over the other voltage gated channels and excellent oral bioavailability. Rhizen intends to move both these compounds to the clinic in 2014.
Details of the presentations:
1. Abstract of the Poster Presentation: “Pre-clinical characterization of RP3128, a novel and potent CRAC channel inhibitor for the treatment of respiratory disorders”
Time and Location- 8 September 2013 between 14.45-16.45 in Room 3.6, at Poster Discussion: New drugs in respiratory medicine, at FIRA BARCELONA, Convention Centre de Gran Via, Barcelona, Spain
2. Abstract of Oral Presentation: “In vitro and in vivo pharmacological profile of RP6503, a novel dual PI3K delta/gamma inhibitor, in pulmonary disease systems”
Time and Location- 11 September 2013 at 8.45 in Room 3.9; Session 8.30-10.30, at the Oral Presentation: Emerging new targets for the treatment of respiratory diseases, at FIRA BARCELONA, Convention Centre de Gran Via, Barcelona, Spain
CLIPS
La Chaux-de-Fonds, Switzerland , Dec. 09, 2015 — Rhizen Pharmaceuticals S.A. announced today that they have entered into an exclusive, worldwide license agreement with Novartis for the development and commercialization of Rhizen’s, inhaled dual PI3K-delta gamma inhibitor and its closely related compounds for various indications.
Under the terms of the agreement, Rhizen will receive an upfront payment and is eligible to receive development, regulatory and sales milestones payments. In addition Rhizen is also eligible to receive tiered royalties on annual nets sales.
The lead compound is a novel, potent, and selective dual PI3K-delta gamma inhibitor with demonstrated anti-inflammatory and immuno-modulatory activity in pre-clinical systems and models representative of respiratory diseases. With a favorable ADME and PK profile and high therapeutic index in animals, the inhaled dual PI3K-delta gamma inhibitor holds promise in the treatment of human airway disorders.
About Rhizen Pharmaceuticals S.A.:
Rhizen Pharmaceuticals is an innovative, clinical-stage biopharmaceutical company focused on the discovery and development of novel therapeutics for the treatment of cancer, immune and metabolic disorders. Since its establishment in 2008, Rhizen has created a diverse pipeline of proprietary drug candidates targeting several cancers and immune associated cellular pathways. Rhizen is headquartered in La-Chaux-de-Fonds, Switzerland. For additional information, please visit Rhizen’s website, http://www.rhizen.com.
SEE
http://www.atsjournals.org/doi/abs/10.1164/ajrccm-conference.2013.187.1_MeetingAbstracts.A3880
| WO2011055215A2 | Nov 3, 2010 | May 12, 2011 | Incozen Therapeutics Pvt. Ltd. | Novel kinase modulators |
| WO2012008302A1 | Jun 28, 2011 | Jan 19, 2012 | National University Corporation Tottori University | Method for preparing novel hipsc by means of mirna introduction |
| WO2012121953A1 | Feb 29, 2012 | Sep 13, 2012 | The Trustees Of Columbia University In The City Of New York | Methods and pharmaceutical compositions for treating lymphoid malignancy |
| WO2012151525A1 | May 4, 2012 | Nov 8, 2012 | Rhizen Pharmaceuticals Sa | Novel compounds as modulators of protein kinases |
| WO2013164801A1 | May 3, 2013 | Nov 7, 2013 | Rhizen Pharmaceuticals Sa | Process for preparation of optically pure and optionally substituted 2- (1 -hydroxy- alkyl) – chromen – 4 – one derivatives and their use in preparing pharmaceuticals |
| US20110118257 | May 19, 2011 | Rhizen Pharmaceuticals Sa | Novel kinase modulators | |
| US20120289496 | May 4, 2012 | Nov 15, 2012 | Rhizen Pharmaceuticals Sa | Novel compounds as modulators of protein kinases |
///////RP 6503, Novartis, develop, commercialize, Rhizen, inhaled dual PI3K-delta gamma inhibitor, PHASE 1, RP-6503
c21c(cccc1O/C(=C(\C2=O)c3cc(ccc3)F)C(C)n4c6ncnc(c6c(n4)c5cc(c(cc5)OC)NS(=O)(=O)C)N)F
CC(C1=C(C(=O)C2=C(O1)C=CC=C2F)C3=CC(=CC=C3)F)N4C5=C(C(=N4)C6=CC(=C(C=C6)OC)NS(=O)(=O)C)C(=NC=N5)N
/////
EDQM adopts revised monograph for WFI allowing non-destillation techniques

In a press release the EDQM has announced that the new monograph draft on Water for Injection (169) had been adopted. Read on to learn more about the production of WFI with membrane systems.
In a press release, the European Pharmacopeia Commission has announced that the revised monograph on Water for Injection (WFI) had been adopted.
According to the revised monograph, it will be allowed in Europe in future to produce WFI with a purification method equivalent to distillation like e.g. reverse osmosis coupled with appropriate techniques. Moreover, the EDQM declares that a notice to the respective supervisory authorities will be required when a “non-distillation” technology is used for the production of WFI. Besides, the EDQM points out that it is not only a matter of equivalence of a specification but rather the robustness of the purification of WFI. Therefore, Annex 1, which is currently under revision, will also include requirements with…
View original post 90 more words
QP Education and Qualification – What is needed?
We are frequently asked about the educational requirements in order to become a Qualified Person in Europe. Comprehensive educational modules are offered, especially in the UK. These training courses contain different topics like pharmaceutical law, Microbiology, Quality Management etc and require the trainee to take part in multiple courses over an extended period. But is this needed to become a QP in Europe?
Read more about QP education and qualification.
We are frequently asked about the educational requirements in order to become a Qualified Person in Europe. Comprehensive educational modules are offered, especially in the UK. These training courses contain different topics like pharmaceutical law, Microbiology, Quality Management etc and require the trainee to take part in multiple courses over an extended period. But is this needed to become a QP in Europe?
The answer comes in two parts.
First: If you are located in the UK then those…
View original post 445 more words
IACS -9571
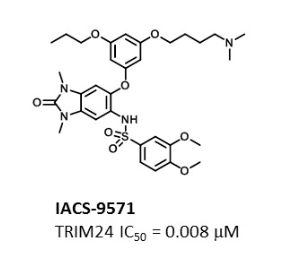

IACS-9571
TRIM24/BRPF1 bromodomain inhibitor
IACS-9571; IACS 9571; IACS9571.
| Molecular Formula: | C32H42N4O8S |
|---|---|
| Molecular Weight: | 642.76288 g/mol |
N-[6-[3-[4-(dimethylamino)butoxy]-5-propoxyphenoxy]-1,3-dimethyl-2-oxobenzimidazol-5-yl]-3,4-dimethoxybenzenesulfonamide
BOARD OF REGENTS, UNIVERSITY OF TEXAS SYSTEM
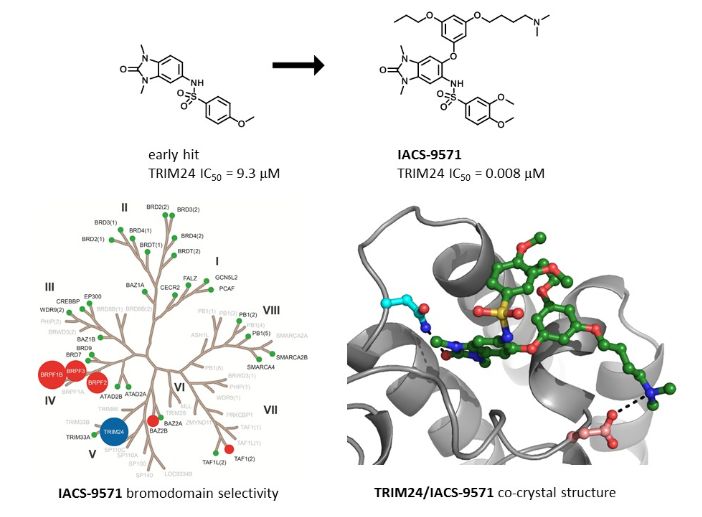
IACS-9571 is a potent and selective inhibitor TRIM24 and BRPF1. The bromodomain containing proteins TRIM24 (Tripartite motif containing protein 24) and BRPF1 (bromodomain and PHD finger containing protein 1) are involved in the epigenetic regulation of gene expression and have been implicated in human cancer. Overexpression of TRIM24 correlates with poor patient prognosis and BRPF1 is a scaffolding protein required for the assembly of histone acetyltransferase complexes, where the gene of MOZ (monocytic leukemia zinc finger protein) was first identified as a recurrent fusion partner in leukemia patients (8p11 chromosomal rearrangements). IACS-9571 has low nanomolar affinities for TRIM24 and BRPF1 (ITC Kd = 31 nM and 14 nM, respectively). With its excellent cellular potency (EC50 = 50 nM) and favorable pharmacokinetic properties (F = 29%), IACS-9571 is a high-quality chemical probe for the evaluation of TRIM24 and/or BRPF1 bromodomain function in vitro and in vivo. (J Med Chem. 2015 Jun 10. [Epub ahead of print] )

PAPER
http://pubs.acs.org/doi/abs/10.1021/acs.jmedchem.5b00405
Structure-Guided Design of IACS-9571, a Selective High-Affinity Dual TRIM24-BRPF1 Bromodomain Inhibitor
†Institute for Applied Cancer Science, and ‡Core for Biomolecular Structure and Function, The University of Texas MD Anderson Cancer Center, 1881 East Road, Unit 1956, Houston, Texas 77054, United States
§ Department of Epigenetics and Molecular Carcinogenesis, The University of Texas MD Anderson Cancer Center,
1515 Holcombe Boulevard
, Houston, Texas 77030, United States
J. Med. Chem., 2016, 59 (4), pp 1440–1454
DOI: 10.1021/acs.jmedchem.5b00405
Publication Date (Web): June 10, 2015
Copyright © 2015 American Chemical Society

The bromodomain containing proteins TRIM24 (tripartite motif containing protein 24) and BRPF1 (bromodomain and PHD finger containing protein 1) are involved in the epigenetic regulation of gene expression and have been implicated in human cancer. Overexpression of TRIM24 correlates with poor patient prognosis, and BRPF1 is a scaffolding protein required for the assembly of histone acetyltransferase complexes, where the gene of MOZ (monocytic leukemia zinc finger protein) was first identified as a recurrent fusion partner in leukemia patients (8p11 chromosomal rearrangements). Here, we present the structure guided development of a series of N,N-dimethylbenzimidazolone bromodomain inhibitors through the iterative use of X-ray cocrystal structures. A unique binding mode enabled the design of a potent and selective inhibitor 8i (IACS-9571) with low nanomolar affinities for TRIM24 and BRPF1 (ITC Kd = 31 nM and ITC Kd = 14 nM, respectively). With its excellent cellular potency (EC50 = 50 nM) and favorable pharmacokinetic properties (F = 29%), 8i is a high-quality chemical probe for the evaluation of TRIM24 and/or BRPF1 bromodomain function in vitro and in vivo.
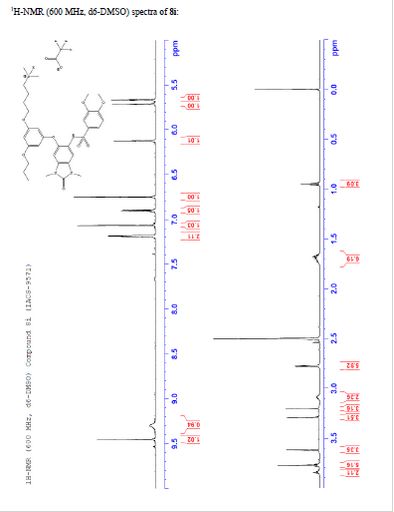
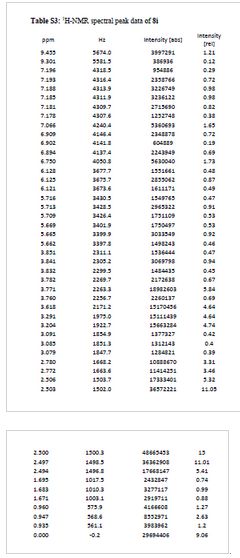
TFA salt of 8i (106 mg, 57%) as a white solid. 1H NMR (600 MHz, DMSO-d6) δ 9.46 (s, 1H), 9.30 (br-s, 1H), 7.19 (m, 2H), 7.07 (s, 1H), 6.90 (d, J = 9.0 Hz, 1H), 6.75 (s, 1H), 6.13 (t, J = 2.2 Hz, 1H), 5.71 (t, J = 2.0 Hz, 1H), 5.67 (t, J = 2.0 Hz, 1H), 3.84 (t, J = 5.9 Hz, 2H), 3.77 (m, 5H), 3.62 (s, 3H), 3.29 (s, 3H), 3.20 (s, 3H), 3.12–3.05 (m, 2H), 2.78 (d, J = 4.7 Hz, 6H), 1.77–1.63 (m, 6H), 0.95 (t, J = 7.3 Hz, 3H). 13C NMR (600 MHz, DMSO-d6) δ 160.3, 160.0, 159.3, 154.1, 152.0, 148.4, 143.9, 131.8, 128.2, 126.0, 121.9, 120.5, 110.4, 109.4, 106.4, 100.6, 95.9, 95.8, 95.2, 68.9, 66.7, 56.3, 55.6, 55.4, 42.1, 27.1, 27.0, 25.6, 21.9, 20.7, 10.4. MS (ESI) m/z 644 [M + H]+.
NMR
IACS -9571
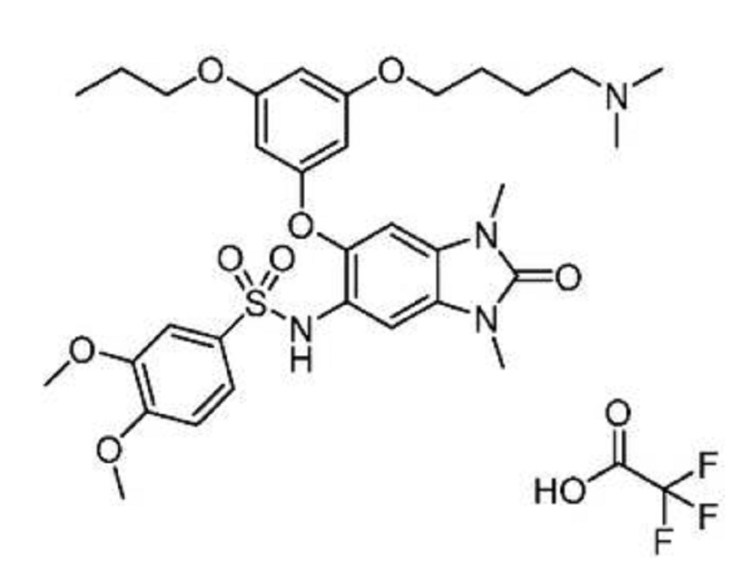
N-(6-(3-(4-(dimethylamino)butoxy)-5- propoxyphenoxy)-l,3-dimethyl-2-oxo-2,3-dihydro-lH-benzo[d]imidazol-5-yl)-3,4- dimethoxybenzenesulfonamide 2,2,2-trifluoroacetate
 CLICK ON IMAGE
ABSTRACT
251st ACS National Meeting & Exposition
13–17 March 2016
San Diego, United States
MEDI 5
Discovery and development of a potent dual TRIM24/BRPF1 bromodomain inhibitor, IACS -9571, using structure- based drug design Wylie S. Palmer 1 , wpalmer@mdanderson.org, Guillaume Poncet -Montagne 1 , Gang Liu 1 , Alessia Petrocchi 1 , N aphtali Reyna 1 , Govindan Subramanian 1 , Jay Theroff 1 , Maria Kost -Alimova 1 , Jennifer Bardenhagen 1 , Elisabetta Leo 1 , Hannah Sheppard 1 , Trang Tieu 1 , Shi Xi 1 , Yanai Zhan 1 , Shuping Zhao 1 , Michelle Barton 2 , Giulio Draetta 1 , Carlo Toniatti 1 , Philip Jones 1 , Mary Ge ck Do 1 , Jannik Andersen 1 . (1) Institute for Applied Cancer Science, The University of Texas, MD Anderson Cancer Center, Houston, Texas, United States (2) Department of Epigenetics and Molecular Carcinogenesis, The University of Texas, MD Anderson Cancer Center, Houston, Texas, United States
Bromodomains are an important class of chromatin remodeling proteins that recognize acetylated lysine residues on histone tails. As epigenetic targets they regulate gene transcription and offer a new way to treat diseas es, particularly in inflammation and oncology. The bromodomain and extra- terminal (BET) family has emerged as an important and druggable example of this class of proteins with the successful entry of small- molecule inhibitors into the clinic. Other families of bromodomains are only starting to be explored, such as the Tripartite Motif -containing 24 protein (TRIM24) and bromodomain- PHD finger protein 1 (BRPF1). Both proteins contain a dual PHD -bromo motif which have a role in recognizing specific histone mar ks. TRIM24 recognizes the dual histone marks of unmodified H3K4 and acetylated- H3K23 within the same histone tail. TRIM24 is a potent co- activator of ER -alpha and overexpression of TRIM24 has been linked to poor survival rates in breast cancer patients.
This presentation will describe the structure guided development of a series of N,N- dimethyl -benzimidazolones through the iterative use of X -ray cocrystal structures. A unique binding mode enabled the design of a potent and selective inhibitor (IACS -9571) with low nanomolar affinities for TRIM24 and BRPF1 (ITC Kd = 31 nM and ITC Kd = 14 nM, respectively). With its excellent cellular potency (EC 50 = 50 nM) and favorable pharmacokinetic properties, IACS -9571 is a high- quality chemical probe for the evaluation of TRIM24 and/or BRPF1 bromodomain function in vitro and in vivo
PATENT
WO-2016033416-A1
Synthesis of Intermediates:
N-(6-bromo-l ,3-dimethyl-2-oxo-2,3-dihydro-lH-benzo[d]imidazol-5-yl)-2,2,2- trifluoroacetamide (Intermediate 1):
Step 1 : 5-nitro-lH-benzo[d]imidazol-2(3H)-one:
To a 0 °C solution of 4-nitrobenzene- 1 ,2-diamine (44 g, 285 mmol) in 80 mL of DMF was added l, l’-carbonyldiimidazole (70 g, 428 mmol). The reaction mixture was stirred at RT for 4 h, then water (250 mL) was added. The resulting suspension was filtered, and the collected solids were washed with water (200 mL) and dried to give 5-nitro-lH- benzo[d]imidazol-2(3H)-one as a yellow solid (45 g, 88%). MS (ES+) C7H5N3O3 requires: 179, found: 180 [M+H]+.
Step 2: l,3-dimethyl-5-nitro-lH-benzo[d]imidazol-2(3H)-one:
To a solution of 5-nitro-lH-benzo[d]imidazol-2(3H)-one (55 g, 309 mmol) in 150 mL of DMF was added K2CO3 (85 g, 618 mmol), the reaction mixture was cooled to 0 °C, then iodomethane (109 g, 772 mmol) was slowly added. The reaction mixture was stirred at RT overnight, then water was added to the reaction mixture. The resulting suspension was filtered and the collected solids were washed with water (200 mL) and dried to give 1,3- dimethyl-5-nitro-lH-benzo[d]imidazol-2(3H)-one as a yellow solid (55 g, 86%). MS (ES+) C9H9N3O3 requires: 207, found: 208 [M+H] +.
Step 3: 5-amino-l,3-dimethyl-lH-benzo[d]imidazol-2(3H)-one:
To a solution of l,3-dimethyl-5-nitro-lH-benzo[d]imidazol-2(3H)-one (50 g, 240 mmol) in 200 mL of EtOAc under an inert atmosphere was added 10% palladium on activated carbon (5 g, 24 mmol). The reaction mixture was then charged with hydrogen and stirred at RT under an ¾ atmosphere overnight. The reaction mixture was filtered through a pad of celite then concentrated to give 5-amino-l,3-dimethyl-lH-benzo[d]imidazol-2(3H)- one as a yellow solid (32 g, 68%). MS (ES+) C9H11N3O requires: 177, found: 178 [M+H]+.
Step 4: 5-amino-6-bromo-l ,3-dimethyl-lH-benzo[d]imidazol-2(3H)-one:
To a 0 °C solution of 5-amino-l ,3-dimethyl-lH-benzo[d]imidazol-2(3H)-one (4 g, 22.6 mmol) in 25 mL of CHCI3 and 25 mL of AcOH was slowly added drop wise bromine (3.5 g, 22.6mmol). The mixture was stirred at RT for 30 min, then concentrated and purified by silica gel chromatography (1 : 1 EtOAc/ hexanes) to afford 5-amino-6-bromo-l ,3-dimethyl- lH-benzo[d]imidazol-2(3H)-one as a yellow solid (3.2 g, 69%). MS (ES+) C9HioBrN30 requires: 256, found: 257 [M+H]+.
Step 5: N-(6-bromo-l ,3-dimethyl-2-oxo-2,3-dihydro-lH-benzo[d]imidazol-5-yl)-2,2,2- trifluoroacetamide:
To a 0 °C solution of 5-amino-6-bromo-l ,3-dimethyl-lH-benzo[d]imidazol- 2(3H)-one (1.50 g, 5.9 mmol) in DCM (45 ml) was added DMAP (72 mg, 0.59 mmol), triethylamine (1.63 ml, 11.7 mmol) and trifluoroacetic anhydride (0.91 ml, 6.4 mmol). The reaction mixture was stirred for 2 h and warmed to RT. The reaction mixture was then quenched with water and the organic phase was washed with brine, dried over sodium sulfate, filtered and concentrated to give N-(6-bromo-l,3-dimethyl-2-oxo-2,3-dihydro-lH- benzo[d]imidazol-5-yl)-2,2,2-trifluoroacetamide (Intermediate 1) as a yellow solid (2.20 g, 100%). MS (ES+) CiiH9BrF3N302 requires: 352, found 353 [M+H]+.
5-amino-6-(3-hydroxyphenoxy)-l,3-dimethyl-lH-benzo[d]imidazol-2(3H)-one (Intermediate 2, Route A):
To a mixture of 5-amino-6-(3-(benzyloxy)phenoxy)-l,3-dimethyl-lH- benzo[d]imidazol-2(3H)-one (400 mg, 1.07 mmol) in DCM (20 mL) at -78 °C was added tribromoborane (5.3 mL, 5.3 mmol). The mixture was warmed up to room temperature gradually, then quenched by methanol dropwise, concentrated, and purified by column chromatography (20-100% EtOAc/hexanes and then 0-40% methanol/EtOAc) to give 5- amino-6-(3-hydroxyphenoxy)-l,3-dimethyl-lH-benzo[d]imidazol-2(3H)-one as a solid (240 mg, 79%). MS (ES+) C15H15N3O3 requires: 285, found: 286 [M+H]+.
5-amino-6-(3-hydroxyphenoxy)-l,3-dimethyl-lH-benzo[d]imidazol-2(3H)-one (Intermediate 2, Route B):
Step 2
Step 1: 3-[(ieri-butyldimethylsilyl)oxy]phenol:
A mixture of lH-imidazole (2.25 g, 33.1 mmol), ieri-butylchlorodimethylsilane (3.83 g, 25.4 mmol) and resorcinol (5.6 g, 51 mmol) in THF (30 ml) was stirred at 80 °C for 5 h. The resulting suspension of the cooled reaction mixture was filtered and the collected filtrate was concentrated and purified by silica-gel chromatography (20:80 to 0:100, EtOAc/hexanes) to give 3-((ieri-butyldimethylsilyl)oxy)phenol (2.78 g, 49%). MS (ES+) C12H20O2S1 requires: 224, found 225 [M+H]+.
Step 2: 5-amino-6-(3-((ier^butyldimethylsilyl)oxy)phenoxy)-l ,3-dimethyl-lH- benzo[d]imidazol-2(3H)-one:
A mixture of 3-((ieri-butyldimethylsilyl)oxy)phenol (1.39 g, 6.20 mmol), quinolin-8-ol (79 mg, 0.55 mmol), copper(I) chloride (20 mg, 0.21 mmol), potassium phosphate (526 mg, 2.48 mmol) and 5-amino-6-bromo-l ,3-dimethyl-lH-benzo[d]imidazol- 2(3H)-one (529 mg, 2.07 mmol) in diglyme (20 ml) in a 100 mL round-bottom flask was degassed under a nitrogen atmosphere and heated to 120 °C for 24 h. To the cooled reaction mixture was added silica gel, stirred for 2 min, then the mixture was filtered through a pad of silica gel. The collected filtrate was concentrated and purified by column chromatography (20:80 to 0: 100, EtOAc/hexanesthen 0: 100 to 40:60, MeOH/EtOAc) to give 5-amino-6-(3- ((ieri-butyldimethylsilyl)oxy)phenoxy)-l,3-dimethyl-lH-benzo[d]imidazol-2(3H)-one (521 mg, 63%). MS (ES+) C21H29N3O3S1 requires: 399, found 400 [M+H]+.
Step 3: 5-amino-6-(3-hydroxyphenoxy)-l,3-dimethyl-lH-benzo[d]imidazol-2(3H)-one:
To a 0 °C solution of 5-amino-6-(3-((ieri-butyldimethylsilyl)oxy)phenoxy)-l,3- dimethyl-lH-benzo[d]imidazol-2(3H)-one (623 mg, 1.56 mmol) in THF was added a solution of ieira-butylammonium fluoride (0.90 mL, 3.1 mmol) in THF, the reaction mixture was allowed to warm up to RT and then stirred for 1-2 h. The reaction mixture was quenched with 1 M hydrogen chloride (0.10 mL, 3.1 mmol) and then partitioned between EtOAc and water. The seperated organic layer was washed with water twice, then concentrated and purified by column chromatography (20-80% EtOAc/hexanes and 0-40% MeOH/DCM) to give 5-amino-6-(3-hydroxyphenoxy)-l ,3-dimethyl-lH-benzo[d]imidazol-2(3H)-one (120 mg, 27%) as a solid. MS (ES+) C15H15N3O3 requires: 285, found 286 [M+H]+.
EXAMPLE 10: N-(6-(3-(4-(dimethylamino)butoxy)-5-propoxyphenoxy)-l,3-dimethyl-2- oxo-2,3-dihydro-lH-benzo[d]imidazol-5-yl)-3,4-dimethoxybenzenesulfonamide 2,2,2-
To a solution of N-(6-(3-(4-aminobutoxy)-5-propoxyphenoxy)-l ,3-dimethyl-2- oxo-2,3-dihydro-lH-benzo[d]imidazol-5-yl)-3,4-dimethoxybenzenesulfonamide 2,2,2- trifluoroacetate (180 mg, 0.247 mmol) in methanol (3.0 ml) was added triethylamine (0.034 ml, 0.25 mmol), acetic acid (0.028 ml, 0.49 mmol), formaldehyde (0.054 ml, 2.0 mmol), and sodium triacetoxyborohydride (131 mg, 0.618 mmol). The reaction mixture was stirred at room temperature and checked by LCMS every 30 minutes. After 3 h the reaction was complete by LCMS. The reaction was quenched with a few drops of TFA and concentrated under reduced pressure. The residue was purified by prep-HPLC using a gradient of 20-60% ACN/water containing 0.1% TFA to afford N-(6-(3-(4-(dimethylamino)butoxy)-5- propoxyphenoxy)-l,3-dimethyl-2-oxo-2,3-dihydro-lH-benzo[d]imidazol-5-yl)-3,4- dimethoxybenzenesulfonamide 2,2,2-trifluoroacetate (106 mg, 57%) as a white solid. MS (ES+) C32H42N4O8S requires: 642, found 643 [M+H]+. ¾ NMR (600 MHz, DMSO-ifc) δ 9.46 (s, 1H), 9.30 (br-s, 1H), 7.19 (m, 2H), 7.07 (s, 1H), 6.90 (d, 7 = 9.0 Hz, 1H), 6.75 (s, 1H), 6.13 (t, 7 = 2.2 Hz, 1H), 5.71 (t, J = 2.0 Hz, 1H), 5.67 (t, J = 2.0 Hz, 1H), 3.84 (t, 7 = 5.9 Hz, 2H), 3.77 (m, 5H), 3.62 (s, 3H), 3.29 (s, 3H), 3.20 (s, 3H), 3.12-3.05 (m, 2H), 2.78 (d, 7 = 4.7 Hz, 6H), 1.77-1.63 (m, 6H), 0.95 (t, 7 = 7.3 Hz, 3H)
References
1: Palmer WS, Poncet-Montange G, Liu G, Petrocchi A, Reyna N, Subramanian G, Theroff J, Yau A, Kost-Alimova M, Bardenhagen JP, Leo E, Shepard HE, Tieu TN, Shi X, Zhan Y, Zhao S, Draetta G, Toniatti C, Jones P, Geck Do M, Andersen JN. Structure-Guided Design of IACS-9571, a Selective High-Affinity Dual TRIM24-BRPF1 Bromodomain Inhibitor. J Med Chem. 2015 Jun 10. [Epub ahead of print] PubMed PMID: 26061247.
US-20160060260-A1
Institute for Applied Cancer Science, The University of Texas, MD Anderson Cancer Center, Houston, Texas, United States

The University of Texas MD Anderson Cancer Center | University of Texas System

The new Institute for Applied Cancer Science will be located at the south campus of M.D.
Draetta arrived at MD Anderson in 2011 to direct the Institute for Applied Cancer Science. He oversees the moon shots platforms
Department of Epigenetics and Molecular Carcinogenesis, The University of Texas, MD Anderson Cancer Center, Houston, Texas, United States

///////IACS-9571, TRIM24, BRPF1 bromodomain inhibitor, IACS-9571, IACS 9571, IACS9571, BOARD OF REGENTS, UNIVERSITY OF TEXAS SYSTEM
CAS BASE 1800477-30-8
CAS OF 1:1 TRIFLUOROACETATE 1883598-69-3
c1(cc(cc(c1)OCCC)Oc3cc2N(C(N(c2cc3NS(=O)(=O)c4cc(c(cc4)OC)OC)C)=O)C)OCCCCN(C)C
CCCOC1=CC(=CC(=C1)OC2=C(C=C3C(=C2)N(C(=O)N3C)C)NS(=O)(=O)C4=CC(=C(C=C4)OC)OC)OCCCCN(C)C
TFA salt of 8i (106 mg, 57%) as a white solid. 1H NMR (600 MHz, DMSO-d6) δ 9.46 (s, 1H), 9.30 (br-s, 1H), 7.19 (m, 2H), 7.07 (s, 1H), 6.90 (d, J = 9.0 Hz, 1H), 6.75 (s, 1H), 6.13 (t, J = 2.2 Hz, 1H), 5.71 (t, J = 2.0 Hz, 1H), 5.67 (t, J = 2.0 Hz, 1H), 3.84 (t, J = 5.9 Hz, 2H), 3.77 (m, 5H), 3.62 (s, 3H), 3.29 (s, 3H), 3.20 (s, 3H), 3.12–3.05 (m, 2H), 2.78 (d, J = 4.7 Hz, 6H), 1.77–1.63 (m, 6H), 0.95 (t, J = 7.3 Hz, 3H). 13C NMR (600 MHz, DMSO-d6) δ 160.3, 160.0, 159.3, 154.1, 152.0, 148.4, 143.9, 131.8, 128.2, 126.0, 121.9, 120.5, 110.4, 109.4, 106.4, 100.6, 95.9, 95.8, 95.2, 68.9, 66.7, 56.3, 55.6, 55.4, 42.1, 27.1, 27.0, 25.6, 21.9, 20.7, 10.4. MS (ESI) m/z 644 [M + H]+.
HELP, Need one time help to pay 10 year concessional subscription to this, your favorite blog to WordPress
Just One viewer please come forward
Dear Kind Viewer’s
WordPress is kind to me and negotiated a one time 10 year concessional subscription of 260 US dollars…….https://newdrugapprovals.org/
I need one time help to pay this one time 10 year concessional subscription to our favorite blog.
This is done to keep this blog running even after my death.
Currently I am paying 99 US Dollars per annum
email me
amcrasto@gmail.com
call +919323115463
Paypal will work for me via email request to you by me, Indian govt does not allow automatic transfer via paypal buttons on the blog.
email me at amcrasto@gmail.com and tell me amount, i will request you via paypal

DR ANTHONY CRASTO
LIONEL MY SON, MY MOTIVATION
.
He was only in first standard in school when I was hit by a deadly one in a million spine stroke called acute transverse mylitis, it made me 90% paralysed…
View original post 226 more words


