New Drug Approvals

Home » Uncategorized (Page 78)

Category Archives: Uncategorized

DRUG APPROVALS BY DR ANTHONY MELVIN CRASTO .....FOR BLOG HOME CLICK HERE

Blog Stats

  • 5,049,583 hits

Flag and hits

Flag Counter

Enter your email address to follow this blog and receive notifications of new posts by email.

Join 37.7K other subscribers
Follow New Drug Approvals on WordPress.com

Archives

Categories

Recent Posts

Flag Counter

ORGANIC SPECTROSCOPY

Read all about Organic Spectroscopy on ORGANIC SPECTROSCOPY INTERNATIONAL 

Enter your email address to follow this blog and receive notifications of new posts by email.

Join 37.7K other subscribers
DR ANTHONY MELVIN CRASTO Ph.D

DR ANTHONY MELVIN CRASTO Ph.D

DR ANTHONY MELVIN CRASTO, Born in Mumbai in 1964 and graduated from Mumbai University, Completed his Ph.D from ICT, 1991,Matunga, Mumbai, India, in Organic Chemistry, The thesis topic was Synthesis of Novel Pyrethroid Analogues, Currently he is working with AFRICURE PHARMA, ROW2TECH, NIPER-G, Department of Pharmaceuticals, Ministry of Chemicals and Fertilizers, Govt. of India as ADVISOR, earlier assignment was with GLENMARK LIFE SCIENCES LTD, as CONSUlTANT, Retired from GLENMARK in Jan2022 Research Centre as Principal Scientist, Process Research (bulk actives) at Mahape, Navi Mumbai, India. Total Industry exp 32 plus yrs, Prior to joining Glenmark, he has worked with major multinationals like Hoechst Marion Roussel, now Sanofi, Searle India Ltd, now RPG lifesciences, etc. He has worked with notable scientists like Dr K Nagarajan, Dr Ralph Stapel, Prof S Seshadri, etc, He did custom synthesis for major multinationals in his career like BASF, Novartis, Sanofi, etc., He has worked in Discovery, Natural products, Bulk drugs, Generics, Intermediates, Fine chemicals, Neutraceuticals, GMP, Scaleups, etc, he is now helping millions, has 9 million plus hits on Google on all Organic chemistry websites. His friends call him Open superstar worlddrugtracker. His New Drug Approvals, Green Chemistry International, All about drugs, Eurekamoments, Organic spectroscopy international, etc in organic chemistry are some most read blogs He has hands on experience in initiation and developing novel routes for drug molecules and implementation them on commercial scale over a 32 PLUS year tenure till date Feb 2023, Around 35 plus products in his career. He has good knowledge of IPM, GMP, Regulatory aspects, he has several International patents published worldwide . He has good proficiency in Technology transfer, Spectroscopy, Stereochemistry, Synthesis, Polymorphism etc., He suffered a paralytic stroke/ Acute Transverse mylitis in Dec 2007 and is 90 %Paralysed, He is bound to a wheelchair, this seems to have injected feul in him to help chemists all around the world, he is more active than before and is pushing boundaries, He has 100 million plus hits on Google, 2.5 lakh plus connections on all networking sites, 100 Lakh plus views on dozen plus blogs, 227 countries, 7 continents, He makes himself available to all, contact him on +91 9323115463, email amcrasto@gmail.com, Twitter, @amcrasto , He lives and will die for his family, 90% paralysis cannot kill his soul., Notably he has 38 lakh plus views on New Drug Approvals Blog in 227 countries......https://newdrugapprovals.wordpress.com/ , He appreciates the help he gets from one and all, Friends, Family, Glenmark, Readers, Wellwishers, Doctors, Drug authorities, His Contacts, Physiotherapist, etc He has total of 32 International and Indian awards

Verified Services

View Full Profile →

Archives

Categories

Flag Counter

ATR 101


PD 132301-2.png

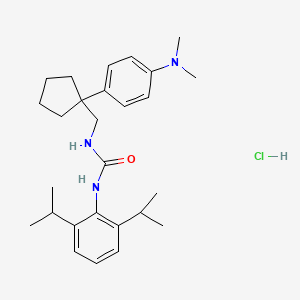
N-(2,6-bis(1-methylethyl)phenyl)-N’-((1-(4-(dimethylamino)phenyl)cyclopentyl) methyl)urea hydrochloride

N-(2,6-BIS(l-METHYLETHYL)PHENYL)-N’-((l-(4- (DIMETHYLAMINO)PHENYL)CYCLOPENTYL)METHYL)UREA

ATR-101; ATR 101; ATR101; PD132301-2; PD-132301-2; PD 132301-2; PD132301; PD-132301; PD 132301.

IUPAC/Chemical Name: 1-(2,6-diisopropylphenyl)-3-((1-(4-(dimethylamino)phenyl)cyclopentyl)methyl)urea hydrochloride

ATR-101 HCl
CAS#: 133825-81-7 (ATR-101 HCl); 133825-80-6 (ATR-101).

Molecular Formula: C27H40ClN3O
Molecular Weight: 458.079 g/mol
The Regents Of The University Of Michigan, Atterocor, Inc.

Millendo Therapeutics is developing ATR-101, an ACAT1 inhibitor, for treating adrenal cancers including adrenocortical cancer and congenital adrenal hyperplasia.


ATR-101, also known as PD-132301 (a free base) or PD-132301-2 (a HCl salt), is in clinical development for the treatment of adrenocortical carcinoma (ACC). ATR-101 is a selective inhibitor of ACAT1 (acyl coenzyme A:cholesterol acyltransferase). ACAT1 catalyzes cholesterol ester formation and, in the adrenals, is particularly important in creating a reservoir of substrate for steroid biosynthesis. ATR-101 is uniquely distributed to adrenal tissues and inhibition of adrenal ACAT1 by ATR-101 disrupts steroidogenesis and leads to selective apoptosis of steroid producing adrenocortical-derived cells. Similar effects have been seen in the human ACC cell line, H295R. ATR-101 has shown pre-clinical efficacy in H295R xenograft mouse models. ACC is an ultra-rare malignancy, occurring in about 2 per million population annually.


ATR-101 (Atterocor, Inc., Ann Arbor, MI, USA) is in clinical development for the treatment of adrenocortical carcinoma (ACC). ATR-101 is a selective inhibitor of ACAT1 (acyl coenzyme A:cholesterol acyltransferase). ACAT1 catalyzes cholesterol ester formation and, in the adrenals, is particularly important in creating a reservoir of substrate for steroid biosynthesis. ATR-101 is uniquely distributed to adrenal tissues and inhibition of adrenal ACAT1 by ATR-101 disrupts steroidogenesis and leads to selective apoptosis of steroid producing adrenocortical-derived cells. Similar effects have been seen in the human ACC cell line, H295R. ATR-101 has shown pre-clinical efficacy in H295R xenograft mouse models. ACC is an ultra-rare malignancy, occurring in about 2 per million population annually. ACC is frequently discovered in Stage 4 and the overall disease survival is approximately 17 months. Tumors often overproduce steroids normally produced in the adrenal cortex. Current therapies are toxic, difficult to administer, and poorly effective. Clinical trial information: NCT01898715.

Adrenocortical carcinoma (ACC) generally has poor prognosis. Existing treatments provide limited benefit for most patients with locally advanced or metastatic tumors. We investigated the mechanisms for the cytotoxicity, xenograft suppression and adrenalytic activity of ATR-101 (PD132301-02), a prospective agent for ACC treatment. Oral ATR-101 administration inhibited the establishment and impeded the growth of ACC-derived H295R cell xenografts in mice. ATR-101 induced H295R cell apoptosis in culture and in xenografts. ATR-101 caused mitochondrial hyperpolarization, reactive oxygen release and ATP depletion within hours after exposure, followed by cytochrome c release, caspase-3 activation, and membrane permeabilization. When combined with ATR-101, lipophilic free radical scavengers suppressed the reactive oxygen release, and glycolytic precursors prevented the ATP depletion, abrogating ATR-101 cytotoxicity. ATR-101 directly inhibited F1F0-ATPase activity and suppressed ATP synthesis in mitochondrial fractions. ATR-101 administration to guinea pigs caused oxidized lipofuscin accumulation in the zona fasciculata layer of the adrenal cortex, implicating reactive oxygen release in the adrenalytic effect of ATR-101. These results support the development of ATR-101 and other adrenalytic compounds for the treatment of ACC.

Company Millendo Therapeutics Inc.
Description Selective inhibitor of sterol O-acyltransferase 1 (SOAT1; ACAT1)
Molecular Target Sterol O-acyltransferase 1 (SOAT1) (ACAT1)


PATENT

WO2013142214

https://www.google.co.in/patents/WO2013142214A1?cl=en

PATENT

WO-2016049518

One such promising agent is N-(2,6-bis( 1 -methylethyl)phenyl)-N’-(( 1 -(4-(dimethyl-amino)phenyl)cyclopentyl)methyl)urea hydrochloride (“ATR-101”). The free base form of ATR-101 has the following chemical structure:

The chemical synthesis of ATR-101 has been previously reported by Trivedi et al. (J. Med. Chem. 37: 1652-1659, 1994). This procedure, however, does not provide for ATR-101 in a form suitable for solid-dosing, particularly with regard to capsule or tablet formation, and does not provide for ATR-101 in high purity.

While significant advances have been made in this field, particularly in the context of ATR-101, there remains a substantial need for improved techniques and products for the oral administration of ATR-101 to patients in need thereof, including patients having ACC and/or other disorders or conditions such as Cushing’s syndrome and congenital adrenal hyperplasia (CAH).


EXAMPLE 1

SYNTHESIS OF SOLID DRUG FORM OF ATR-101

Step 1 : Preparation of Primary Amine 2 from the Nitrile 1

Tetrahyrofuran (THF) and Compound 1 are charged to a reactor vessel and a lithium aluminum hydride (LAH) solution in THF is added slowly. After the addition, the reaction mixture is warmed to 45°C and stirred until in-process HPLC analysis indicates that the reaction is complete. The reaction mixture is cooled to between 0 and 10°C and aqueous NaOH is added slowly while controlling the temperature to between 0 and 10°C. The mixture is then warmed to between 20 and 25°C and any inorganic salts removed by filtration. The solids are then washed with additional THF.

The filtrate is distilled under vacuum. Acetonitrile (MeCN) is added and the distillation continued to reduce the total volume. H20 is added and the solution is cooled to 20°C, and seeded if necessary. Additional water is added to the slurry and cooled to between 0 and 5°C and filtered. The crystallization vessel and filter cake is washed with MeCN and water (1 :2 mixture) and dried under vacuum between 40 to 45°C to produce Compound 2. Typical yield: 85%.

Step 2: Preparation of ATR-101 Free Base

2,6-Diisopropyl aniline hydrochloride (Compound 3) is converted to the corresponding free base by stirring in a mixture of dichloromethane (DCM) and 10% aqueous NaOH. The organic phase is separated and washed with water. The DCM solution containing the aniline free base is concentrated by distillation.

4-dimethylaminopyridine (DMAP) and DCM are charged to a separate reaction vessel. The mixture is cooled and a solution of di-tert-butyl dicarbonate (Boc20) in DCM is slowly added while the temperature is maintained between 0 and 5°C. The aniline free base solution is then slowly added to the reaction vessel. A complete conversion of aniline to the isocyanate is verified by in-process HPLC analysis.

Compound 2 and MeCN are charged to a separate vessel and this solution is cooled to between 0 and 5°C. The isocyanate intermediate solution

(prepared above) is slowly added while the temperature is maintained between 0 and 5°C, and stirred until in-process HPLC indicates that the reaction is complete.

The reaction mixture is distilled under vacuum, and isopropyl alcohol

(IP A) is added and the distillation is continued. The resulting solution is cooled and seeded, if necessary. After crystallization occurs, water is added and the mixture is cooled to between 0 and 5°C, and filtered. The crystallization vessel and filter cake is washed with isopropanol: water (1 : 1) and the product cake is dried under vacuum to yield ATR-101 as the free base. Typical yield: 89 %

Step 3 : Preparation of Solid Drug Form of ATR- 101

The ATR-101 free base is dissolved in acetone and filtered to remove particulates. Additional acetone is used to rinse the dissolution vessel and filter. Concentrated hydrochloric acid (HCl) is added while maintaining the reaction at room temperature. The resultant slurry is filtered and the cake is washed with acetone. The resulting solid is dried under vacuum between 40 and 45°C to obtain the solid drug form of ATR-101. Typical yield: 70-80 %.

EXAMPLE 2

CHARACTERIZATION OF THE SOLID DRUG FORM OF ATR-101

The solid drug form of ATR-101 was analyzed to fully characterize the material and provide proof of structure.

Elemental Analysis

An elemental (CHN) analysis was conducted, in duplicate, of the solid drug form of ATR-101. The results are summarized in Table 1 and are in agreement with the theoretical values calculated for the molecular ATR-101 drug substance formula of C27H39N3O HCl.

Table 1

Chloride Content

The solid drug form of ATR-101 is prepared as its HCl salt. To confirm the chloride content (and the stoichiometry), the hydrochloride salt was analyzed by Ion Chromatography using a validated method. The w/w% result showed 7.8% chloride present. The theoretical value for a mono hydrochloride salt is 7.7%. The experimental result conforms to the theoretical value for the mono-hydrochloride salt.

Mass Spectrometry

Mass spectrometry studies were conducted in accordance with

USP<736> using an AB Sciex API 2000 LC/MS/MS system. The samples were analyzed by electrospray ionization in positive mode. The base peak observed was 422.3 (M+H-HC1), consistent with the parent compound (see Figure 1). Two minor peaks were observed, at 301.3 and 202.3 (uncharacterized fragments). The combined data of the LC/MS and CFIN results support the molecular formula assignment of C27H39N3O and mass of 421.63 g/mol for the free base and C27H39N3O . HCl (mass of 458.09 g/mol) for the mono hydrochloride salt.

Nuclear Magnetic Resonance (NMR) – 1H NMR

The proton NMR spectrum of the solid drug form of ATR-101 was obtained using a Varian Gemini 400 MHz spectrometer and. The sample was dissolved in CD3OD. The resulting proton NMR spectrum is shown in Figure 2.

Two-Dimensional (2D) NMR

The 2D proton NMR spectrum (COSY) shown in Figure 3 confirmed some of the connectivity expected for the solid drug form of ATR-101. In particular the resonance at 1.2 ppm is strongly correlated to the resonance at 3.1. This correlation together with the splitting pattern observed for the peak at 3.1 strongly suggests an isopropyl moiety. Further, the data from these spectra show a strong correlation between each of the broad peaks at 1.6-2.2 ppm, consistent with a cycloalkyl functionality in which no heteroatoms or other non-alkyl substitution is present.

Carbon 13 NMR (13C NMR)

The 100 MHz 13C NMR spectrum of the solid drug form of ATR-101 was obtained using a Varian Gemini 400 MHz spectrometer. The sample was dissolved in CD3OD. The resulting 13C NMR spectrum is shown in Figure 4. The numbering of the carbon atoms for the analysis of the spectrum is shown below, and the interpretation is shown in Table 2. The observed signals are consistent with the structure of ATR-101.

Table 2

Fourier Transform Infrared Spectroscopy (IR)

Infrared (IR) spectroscopy was performed using the soid drug form of ATR-101. The resulting spectrum, shown in Figure 5, is consistent with the structure of ATR-101 drug substance. The major peak assignments are presented in Table_3.

Table 3

EXAMPLE 3

COMPARISON WITH PRIOR ART SYNTHESIS OF ATR-101 (BY TRIVEDI ETAL.. J. MED. CHEM. 37: 1652-1659, 1994)

ATR-101

In this experiment, 10.6 g of ATR-101 was synthesized according to the above procedure, which corresponds to the the procedure set forth in Trivedi et al., J. Med. Chem. 137: 1652-1659, 1994 (hereinafter referred to as the “Trivedi procedure”). The purity of ATR-101 as made by the Trivedi procedure was found to be 94.9%, compared to a purity of 98.3% for ATR-101 obtained by the procedure of Example 1 and as evaluated in Example 2.

Step 1 : Alkylation of p-nitrophenylacetonitrile

52

The initial alkylation reaction was run on 15.0 g scale and, according to the Trivedi procedure, should have given 15.7 g (79%) of product 52. However, several problems occurred, and the yield was much lower than expected (6.0 g, 30% yield), although the purity by 1H NMR and melting point (actual: 71-72°C, reported: 76°C) seemed good. Approximately half way through the addition of 1 ,4-bromobutane and p- nitrophenylacetonitrile to NaH, a black solid precipitated out of the purple solution causing the stirbar in the flask to skip and jump. The rate of stirring had to be monitored throughout the remainder of the addition to maintain a sluggish and inefficient mixing of the solution.

After stirring at ambient temperature overnight to ensure reaction completion, the reaction was worked-up as the procedure indicated. First, excess ether was removed using air bubbling, and the black solid was isolated by filtration. Diethyl ether was then added until all of the solids dissolved to give a clear black solution. However, upon washing the ether solution with 2N HC1, a black amorphous solid precipitated from the solution. There was no note of this black solid in the Trivedi procedure, so the work-up was continued without modification. The black solids ended up in the aqueous washes, or stuck to the seperatory funnel. The remainder of the work-up proceeded as expected, and the hot hexanes extraction of the crude solid resulted in light pink planar crystals.

The procedure was repeated with two changes thought to be responsible for the low yield: the anhydrous solvent (from the bottle) was sieve dried to remove trace water, and the stir bar was replaced with a mechanical stirrer to ensure more even mixing of the solution. The procedure was re-run on 10 g scale, which should have yielded 10.5 g of compound 52. However, despite the changes to the procedure, the resulting product and yield was nearly identical to the first run (4.5 g, 34% yield, 71-72°C melting point).

In an attempt to determine where the bulk of material ended up, the aqueous layer from this reaction was re-extracted with diethyl ether, but only resulted in trace amounts of material. The black solids that formed during the work-up were isolated by filtration, and an NMR was taken of the material. The NMR showed peaks corresponding to compound 52. Presumably, this amorphous black solid that resulted after HC1 formation is the main source of lost material, as there appeared to be several grams of it.

Ste 2: Reduction of Nitro Compound

The conversion of nitro compound 52 to the dimethyl amine 53 was done over two steps: palladium catalyzed hydrogenation of the nitro compound to give the free amine 52b, followed by imine formation & reduction to the dimethylamine 53.

An exploratory small scale reaction was run, using 1/10th of the available material (1.0 g compound 52). The reduction of the nitro compound on the 1 gram scale was very rapid, with hydrogen consumption ceasing after 3-4 hours. A crude NMR of an aliquot of the reaction mixture showed very clean amine (52b). The formaldehyde was added, as well as additional Pd/C, and the hydrogenation was continued. The hydrogen was not consumed as quickly for the imine reduction, and the reaction was still progressing when the vessel was pressurized to 55 psi and left shaking overnight (ca. 16h).

After 16 hours, the pressure in the flask had dropped to 30 psi, indicating that the hydrogenation was still progressing overnight. An aliquot NMR confirmed that the reaction had not proceeded to completion.

On large scale, the nitro reduction proceeded very smoothly, consuming hydrogen at a very rapid rate, and going to completion again within 3-4 hours. The reactor was pressurized to 55 psi and shaken overnight, as indicated in the original procedure, before more Pd/C was added, followed by formaldehyde. Hydrogen consumption was again observed to be very sluggish, so the valve to the hydrogen tank was left open to the vessel, and the reaction was shaken for 24 hours.

After 24 hours of shaking, the valve to the vessel was closed, and a drop of 5 psi was observed over 1 hour, indicating that the reaction had not progressed to completion. TLC also showed several polar products, suggesting that the reaction was only ca. 50% complete. The hydrogenation vessel was pressurized to 55 psi with hydrogen, and the valve again left open for an additional 24 hours of hydrogenation.

After 24 hours, the reaction stopped consuming hydrogen, and the vessel was purged and the contents filtered to remove the palladium catalyst. The work-up was performed similarly to the small scale, and the two reactions were combined prior to purification by column chromatography, giving 5.7g (57.5% yield) of the desired dimethylamine product 53.

Step 3 : Reduction of C ano Compound

A small scale RaNi hydrogenation was done and the test reaction went smoothly. Hydrogen consumption was rapid, and the reaction appeared complete after approximately 2 hours. The consumption of hydrogen had ceased, and TLC indicated that there was no compound 53 remaining. After filtration to remove the Raney Nickel, the reaction completion was confirmed by aliquot NMR.

The remaining material was subjected to reduction using the same conditions, and hydrogen consumption and TLC analysis again indicated reaction completion after 2 hours. The material was filtered and combined with the smaller scale reaction material. After concentration to dryness, the crude yield was found to be 5.5 g (96.5% yield), which was very close to the reported yield (99%>).

Step 4: Formation of Urea Com ound

Urea formation is a straightforward procedure, and the small scale test reaction with the amine 54 (500 mg) being combined with 1.0 equivalent of the

isocyanate in 20 parts ethyl acetate. After stirring for 16 hours, the solution was concentrated to dryness to give a white solid. Crude 1H NMR of the solid confirmed that the spectra matched the reported spectra in the Trivedi procedure.

The remaining material was carried forward to ATR-101 freebase without difficulty, and the lots of product were combined. In an effort to remove the residual ethyl acetate, the solids were dissolved in 10 mL of toluene, followed by concentration under reduced pressure. After drying on high- vacuum, ATR-101 freebase was isolated as a sticky white foam (10.6 g, 99% yield). The 1H NMR of the final product showed trace toluene even after extended drying, and the material was moved on to the HC1 salt formation.

The melting point of the solid was later taken and found to be surprisingly low (50-56°C, expected: 132-133°C). The nature of the solid (oily foam) made the determination of the melting point difficult, but it was judged to be completely melted above 60°C.

Step 5: Formation of HC1 Salt

To the ATR-101 freebase in toluene was added 37% HC1, and a gummy white solid precipitated out immediately. The solution was dried by Dean-Stark apparatus over approximately 3 hours with vigorous stirring and heating (bath temp: 160°C). After drying, the solution was cooled and the fine crystalline solid was isolated by filtration and washed with acetone and diethyl ether. The product ATR-101 was dried until a constant weight was achieved (10.6 g, 92% yield) and fully characterized.

Figure 1 is the LC/MS Mass spectrum of the solid drug form of ATR- 101.

https://patentscope.wipo.int/search/docservice_image_drawings/WO@@@id00000032865992@@@11250297@@@200@@@0@@@000061.tif

Figure 2 is the proton NMR spectrum of the solid drug form of ATR- 101.

Figure 3 is the 2-D 1H NMR spectrum (COSY) of the solid drug form of ATR-101.

Figure 4 is the 13C NMR spectrum of of the solid drug form of ATR- 101.

Figure 5 is the FT-IR spectrum the solid drug form of ATR-101.


Paper

(J. Med. Chem. 37: 1652-1659, 1994

http://pubs.acs.org/doi/abs/10.1021/jm00037a016


Patent ID Date Patent Title
EP0474733 1994-08-31 ANTIHYPERLIPIDEMIC AND ANTIATHEROSCLEROTIC UREA COMPOUNDS.
WO9015048 1990-12-13 ANTIHYPERLIPIDEMIC AND ANTIATHEROSCLEROTIC UREA COMPOUNDS
Patent ID Date Patent Title
US2015087649 2015-03-26 TREATING DISORDERS ASSOCIATED WITH ABERRANT ADRENOCORTICAL CELL BEHAVIOR
US2013267550 2013-10-10 Compounds and Methods for Treating Aberrant Adrenocartical Cell Disorders
EP0858336 2006-12-20 METHOD AND PHARMACEUTICAL COMPOSITION FOR REGULATING LIPID CONCENTRATION
US2005234124 2005-10-20 Carboxyalkylether-ACAT inhibitor combinations
US2004072903 2004-04-15 Carboxyalkylether-acat inhibitors combinations
US6143755 2000-11-07 Pharmaceutical methods of treatment with ACAT inhibitors and HMG-CoA reductase inhibitors
US6124309 2000-09-26 Method and pharmaceutical composition for regulating lipid concentration
US6093719 2000-07-25 Method and pharmaceutical composition for regulating lipid concentration
WO9716184 1997-05-09 METHOD AND PHARMACEUTICAL COMPOSITION FOR REGULATING LIPID CONCENTRATION
EP0474733 1994-08-31 ANTIHYPERLIPIDEMIC AND ANTIATHEROSCLEROTIC UREA COMPOUNDS.

References

1: Wolfgang GH, MacDonald JR, Vernetti LA, Pegg DG, Robertson DG. Biochemical alterations in guinea pig adrenal cortex following administration of PD 132301-2, an inhibitor of acyl-CoA:cholesterol acyltransferase. Life Sci. 1995 Feb 17;56(13):1089-93. PubMed PMID: 9001442.

2: Saxena U, Ferguson E, Newton RS. Acyl-coenzyme A:cholesterol-acyltransferase (ACAT) inhibitors modulate monocyte adhesion to aortic endothelial cells. Atherosclerosis. 1995 Jan 6;112(1):7-17. PubMed PMID: 7772069.

3: Reindel JF, Dominick MA, Bocan TM, Gough AW, McGuire EJ. Toxicologic effects of a novel acyl-CoA:cholesterol acyltransferase inhibitor in cynomolgus monkeys. Toxicol Pathol. 1994 Sep-Oct;22(5):510-8. PubMed PMID: 7899779.

4: Krause BR, Black A, Bousley R, Essenburg A, Cornicelli J, Holmes A, Homan R, Kieft K, Sekerke C, Shaw-Hes MK, et al. Divergent pharmacologic activities of PD 132301-2 and CL 277,082, urea inhibitors of acyl-CoA:cholesterol acyltransferase. J Pharmacol Exp Ther. 1993 Nov;267(2):734-43. PubMed PMID: 8246149.

5: Dominick MA, McGuire EJ, Reindel JF, Bobrowski WF, Bocan TM, Gough AW. Subacute toxicity of a novel inhibitor of acyl-CoA: cholesterol acyltransferase in beagle dogs. Fundam Appl Toxicol. 1993 Feb;20(2):217-24. PubMed PMID: 8383621.

6: Dominick MA, Bobrowski WA, MacDonald JR, Gough AW. Morphogenesis of a zone-specific adrenocortical cytotoxicity in guinea pigs administered PD 132301-2, an inhibitor of acyl-CoA:cholesterol acyltransferase. Toxicol Pathol. 1993;21(1):54-62. PubMed PMID: 8397438.

///////ATR 101, 133825-81-7, ATR-101 HCl,  133825-80-6,  Millendo Therapeutics,  ACAT1 inhibitor, treating adrenal cancers,  adrenocortical cancer,  congenital adrenal hyperplasia, Atterocor, Inc., Ann Arbor, MI, USA

O=C(NCC1(C2=CC=C(N(C)C)C=C2)CCCC1)NC3=C(C(C)C)C=CC=C3C(C)C.[H]Cl
PF-05387552


str1

str1

STR3

CID 50992153.png

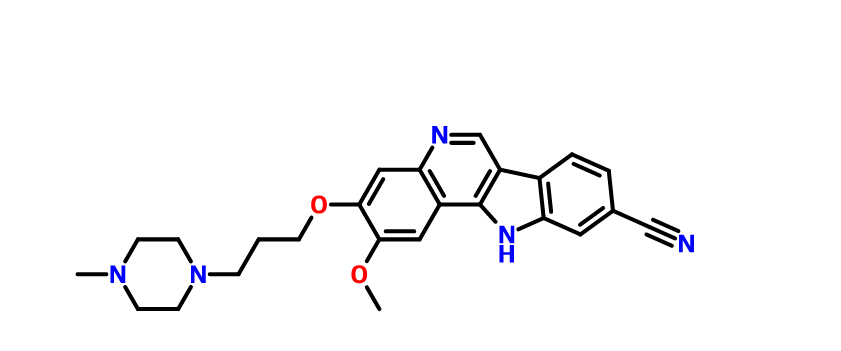
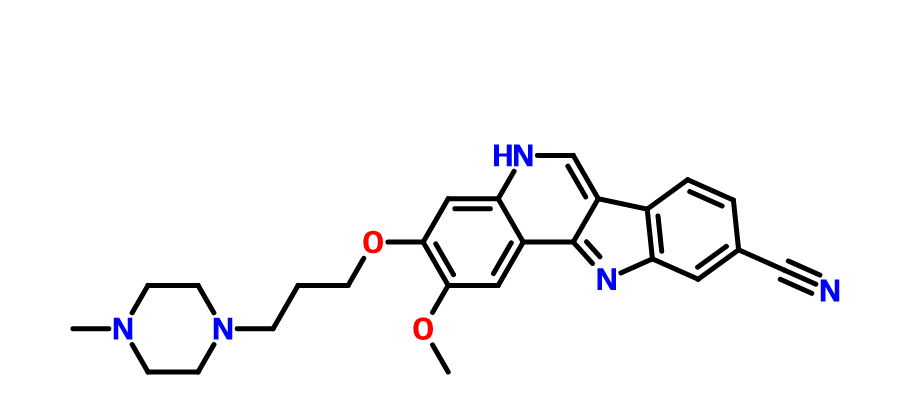
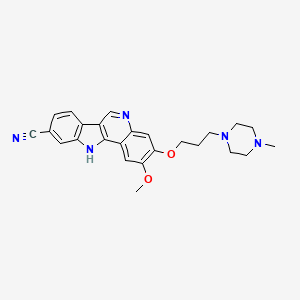
PF-05387552

IRAK4

CAS 1604034-71-0
C25 H27 N5 O2
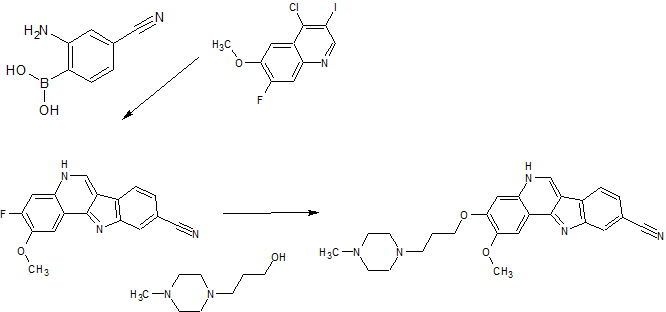
11H-​Indolo[3,​2-​c]​quinoline-​9-​carbonitrile, 2-​methoxy-​3-​[3-​(4-​methyl-​1-​piperazinyl)​propoxy]​-
2-methoxy-3-[3-(4-methylpiperazin-1-yl)propoxy]-11H-indolo[3,2-c]quinoline-9-carbonitrile
Molecular Weight429.51
Molecular Formula: C25H27N5O2
Molecular Weight: 429.51418 g/mol

Synthesis

str1

PAPER

Bioorganic & Medicinal Chemistry Letters (2014), 24(9), 2066-2072

Identification and optimization of indolo[2,3-c]quinoline inhibitors of IRAK4

 a Pfizer Global R&D, 445 Eastern Point Rd., Groton, CT 06340, USA
  • b Pfizer Global R&D, 200 Cambridge Park Dr., Cambridge, MA 02140, USA
  • c Pfizer Global R&D, 87 Cambridgepark Dr., Cambridge, MA 02140, USA
  • d Pfizer Global R&D, 1 Burtt Rd., Andover, MA 01810, USA

http://www.sciencedirect.com/science/article/pii/S0960894X14002832?np=y

Image for unlabelled figure

IRAK4 is responsible for initiating signaling from Toll-like receptors (TLRs) and members of the IL-1/18 receptor family. Kinase-inactive knock-ins and targeted deletions of IRAK4 in mice cause reductions in TLR induced pro-inflammatory cytokines and these mice are resistant to various models of arthritis.

Herein we report the identification and optimization of a series of potent IRAK4 inhibitors. Representative examples from this series showed excellent selectivity over a panel of kinases, including the kinases known to play a role in TLR-mediated signaling. The compounds exhibited low nM potency in LPS- and R848-induced cytokine assays indicating that they are blocking the TLR signaling pathway.

A key compound (26) from this series was profiled in more detail and found to have an excellent pharmaceutical profile as measured by predictive assays such as microsomal stability, TPSA, solubility, and c log P. However, this compound was found to afford poor exposure in mouse upon IP or IV administration. We found that removal of the ionizable solubilizing group (32) led to increased exposure, presumably due to increased permeability. Compounds 26 and 32, when dosed to plasma levels corresponding to ex vivo whole blood potency, were shown to inhibit LPS-induced TNFα in an in vivo murine model.

To our knowledge, this is the first published in vivo demonstration that inhibition of the IRAK4 pathway by a small molecule can recapitulate the phenotype of IRAK4 knockout mice.

L. Nathan TurneyL. Nathan Tumey, Ph.D., Principal Research Scientist, Pfizer Global R&D
REFERENCES

STR3

///////////TLR signaling, Indoloquinoline, IRAK4, Kinase inhibitor, Inflammation, PF-05387552, PF 05387552,  1604034-71-0

N#Cc3ccc4c5cnc2cc(OCCCN1CCN(C)CC1)c(OC)cc2c5nc4c3

Curis and Aurigene’s CA 4948, AU 4948


Curis, Inc.

STR3

CHEMBL3353198.png

Example 13 WO2015104688

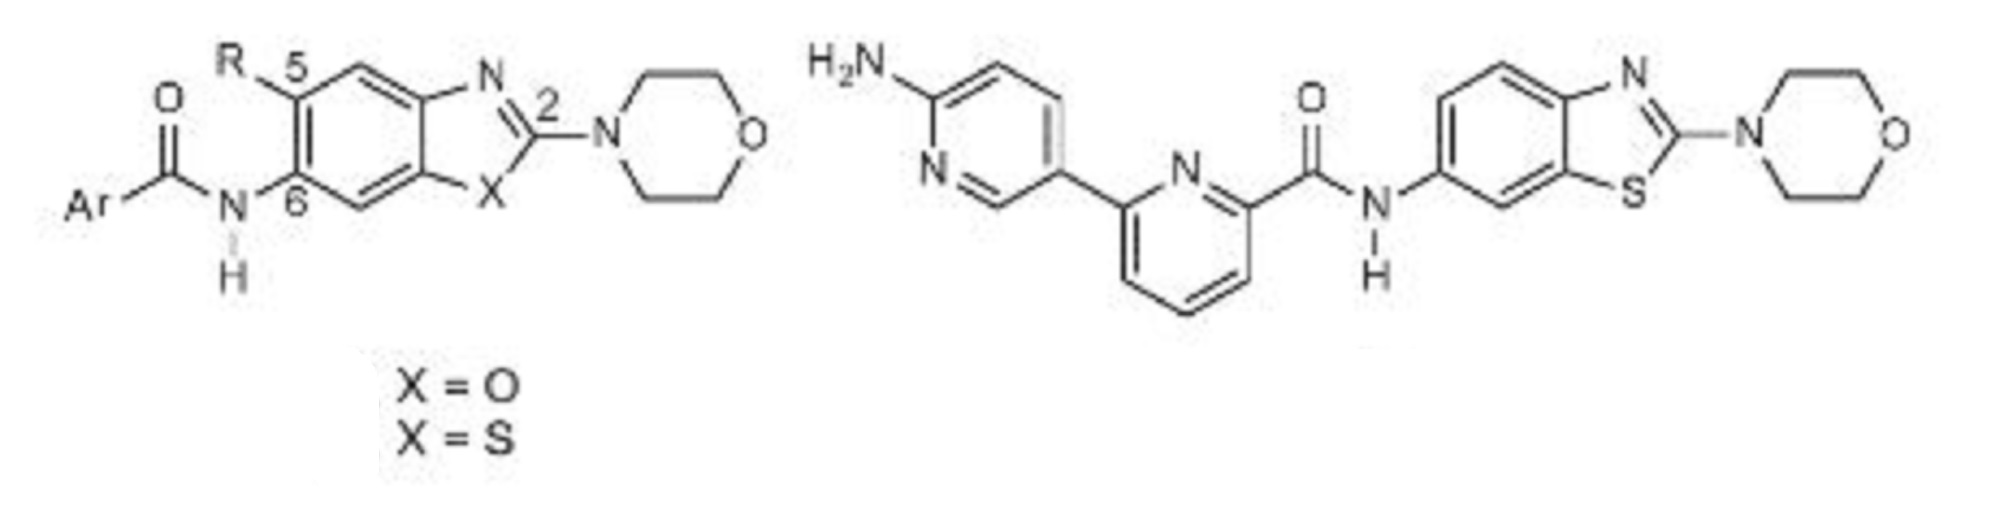
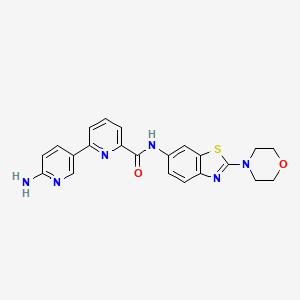
6-(6-aminopyridin-3-yl)-N-(2-morpholin-4-yl-1,3-benzothiazol-6-yl)pyridine-2-carboxamide

Molecular Formula: C22H20N6O2S
Molecular Weight: 432.4982 g/mol
1428335-77-6
[2,​3′-​Bipyridine]​-​6-​carboxamide, 6′-​amino-​N-​[2-​(4-​morpholinyl)​-​6-​benzothiazolyl]​-

PROBABLE STRUCTURE

Example 1 ……..6′-amino-N-(2-morpholinooxazolo[4,5-b]pyridin-6-yl)-[2,3′-bipyridine]-6-carboxamideWO2015104688

STR3

Compound-6:  6′-amino-N-(5-(cyclopropyIamino)-2-morpholinobenzo [d]oxazoI-6-yl)-[2,3′-bipyridine]-6-carboxamide.WO2013042137

PROBABLE CA 4948, AU 4948,AU-4948, CA-4948

STRUCTURE AND SYNTHESIS COMING……..

Company Aurigene Discovery Technologies Ltd.
Description Oral IL-1 receptor-associated kinase 4 (IRAK4) inhibitor
Molecular Target Interleukin-1 receptor-associated kinase 4 (IRAK4)
Mechanism of Action
Therapeutic Modality Small molecule
Latest Stage of Development Preclinical
Standard Indication B cell lymphoma
Indication Details Treat diffuse large B cell lymphoma (DLBCL)
Regulatory Designation
Partner Curis Inc.

Interleukin-1 Receptor Associated Kinase-4 (IRAK-4) is a serine/threonine protein kinase belonging to tyrosine like kinase (TLK) family. IRAK-4 is one of the important signalling components downstream of IL-1/Toll family of receptors (IL-1R, IL-18R, IL-33R, Toll-like receptors). Recent studies have reported occurrence of oncogenic mutations in MYD88 in 30% of ABC diffuse large B cell lymphomas (ABC DLBCL) and 90% of Waldenstrom’s macroglobulinemia (WM). Most of ABC DLBCLs have a single amino acid substitution of proline for the leucine at position 265 (L265P) in the TIR domain of MYD88 protein resulting in constitutive activation of IRAK-4. Thus, IRAK4 is an attractive therapeutic target for the treatment of B-cell lymphomas with activating MYD88 L265P mutation. We have designed, synthesized and tested small molecule IRAK-4 inhibitors based on hits originating from Aurigene’ s compound library. These novel compounds were profiled for IRAK4 kinase inhibition, anti-proliferative activity, kinase selectivity, and drug-like properties. Furthermore, selected compounds were tested in a proliferation assay and pIRAK1 mechanistic assay using ABC-DLBCL cell lines with activating MYD88 L265P mutation, OCI-lLy10 and OCI-lLy3. We have identified a series of novel bicyclic heterocycles as potent inhibitors of IRAK-4. Aurigene Lead compound exhibited potent inhibitory activity for IRAK-4 with an IC50 of 3nM in biochemical assay. Aurigene Lead compound inhibited pIRAK1 levels, and proliferation of OCI-Ly3 and OCI-Ly10 cells with an IC501of 132nM and 52nM respectively. To the best of our knowledge, Aurigene Lead compound represents the most potent IRAK4 inhibitor reported for target modulation and anti-proliferative activity in DLBCL cell lines with activating MYD88 L265P mutation. Aurigene Lead compound has good oral pharmacokinetic profile in mice and has demonstrated excellent pharmacodynamic effect in an in vivo LPS induced TNF-α model with an ED50 of 3.8 mg/Kg in mice. Preliminary in vitro tox studies indicated clean safety profile. Demonstration of efficacy in OCI-lLy10 mouse tumor model is ongoing. In summary, a series of potent IRAK-4 inhibitors belonging to 3 different chemical series have been discovered and are being evaluated for treatment of B-cell lymphomas.

Curis with the option to exclusively license Aurigene’s orally-available small molecule inhibitor of Interleukin-1 receptor-associated kinase 4 (IRAK4) in the precision oncology field. Curis expects to exercise its option to obtain exclusive licenses to both programs and file IND applications for a development candidate from each in 2015.

Recent studies have also shown that alterations of the MYD88 gene lead to dysregulation of its downstream target IRAK4 in a number of hematologic malignancies, including Waldenström’s Macroglobulinemia and a subset of diffuse large B-cell lymphomas, making IRAK4 an attractive target for the treatment of these cancers.

Curis, Inc.

Jan 21, 2015

Curis and Aurigene Announce Collaboration, License and Option Agreement to Discover, Develop and Commercialize Small Molecule Antagonists for Immuno-Oncology and Precision Oncology Targets

— Agreement Provides Curis with Option to Exclusively License Aurigene’s Antagonists for Immuno-Oncology, Including an Antagonist of PD-L1 and Selected Precision Oncology Targets, Including an IRAK4 Kinase Inhibitor —

— Investigational New Drug (IND) Application Filings for Both Initial Collaboration Programs Expected this Year —

— Curis to issue 17.1M shares of its Common Stock as Up-front Consideration —

— Management to Host Conference Call Today at 8:00 a.m. EST —

LEXINGTON, Mass. and BANGALORE, India, Jan. 21, 2015 (GLOBE NEWSWIRE) — Curis, Inc. (Nasdaq:CRIS), a biotechnology company focused on the development and commercialization of innovative drug candidates for the treatment of human cancers, and Aurigene Discovery Technologies Limited, a specialized, discovery stage biotechnology company developing novel therapies to treat cancer and inflammatory diseases, today announced that they have entered into an exclusive collaboration agreement focused on immuno-oncology and selected precision oncology targets. The collaboration provides for inclusion of multiple programs, with Curis having the option to exclusively license compounds once a development candidate is nominated within each respective program. The partnership draws from each company’s respective areas of expertise, with Aurigene having the responsibility for conducting all discovery and preclinical activities, including IND-enabling studies and providing Phase 1 clinical trial supply, and Curis having responsibility for all clinical development, regulatory and commercialization efforts worldwide, excluding India and Russia, for each program for which it exercises an option to obtain a license.

The first two programs under the collaboration are an orally-available small molecule antagonist of programmed death ligand-1 (PD-L1) in the immuno-oncology field and an orally-available small molecule inhibitor of Interleukin-1 receptor-associated kinase 4 (IRAK4) in the precision oncology field. Curis expects to exercise its option to obtain exclusive licenses to both programs and file IND applications for a development candidate from each in 2015.

“We are thrilled to partner with Aurigene in seeking to discover, develop and commercialize small molecule drug candidates generated from Aurigene’s novel technology and we believe that this collaboration represents a true transformation for Curis that positions the company for continued growth in the development and eventual commercialization of cancer drugs,” said Ali Fattaey, Ph.D., President and Chief Executive Officer of Curis. “The multi-year nature of our collaboration means that the parties have the potential to generate a steady pipeline of novel drug candidates in the coming years. Addressing immune checkpoint pathways is now a well validated strategy to treat human cancers and the ability to target PD-1/PD-L1 and other immune checkpoints with orally available small molecule drugs has the potential to be a distinct and major advancement for patients. Recent studies have also shown that alterations of the MYD88 gene lead to dysregulation of its downstream target IRAK4 in a number of hematologic malignancies, including Waldenström’s Macroglobulinemia and a subset of diffuse large B-cell lymphomas, making IRAK4 an attractive target for the treatment of these cancers. We look forward to advancing these programs into clinical development later this year.”

Dr. Fattaey continued, “Aurigene has a long and well-established track record of generating targeted small molecule drug candidates with bio-pharmaceutical collaborators and we have significantly expanded our drug development capabilities as we advance our proprietary drug candidates in currently ongoing clinical studies. We believe that we are well-positioned to advance compounds from this collaboration into clinical development.”

CSN Murthy, Chief Executive Officer of Aurigene, said, “We are excited to enter into this exclusive collaboration with Curis under which we intend to discover and develop a number of drug candidates from our chemistry innovations in the most exciting fields of cancer therapy. This unique collaboration is an opportunity for Aurigene to participate in advancing our discoveries into clinical development and beyond, and mutually align interests as provided for in our agreement.  Our scientists at Aurigene have established a novel strategy to address immune checkpoint targets using small molecule chemical approaches, and have discovered a number of candidates that modulate these checkpoint pathways, including PD-1/PD-L1. We have established a large panel of preclinical tumor models in immunocompetent mice and can show significant in vivo anti-tumor activity using our small molecule PD-L1 antagonists.  We are also in the late stages of selecting a candidate that is a potent and selective inhibitor of the IRAK4 kinase, demonstrating excellent in vivo activity in preclinical tumor models.”

In connection with the transaction, Curis has issued to Aurigene approximately 17.1 million shares of its common stock, or 19.9% of its outstanding common stock immediately prior to the transaction, in partial consideration for the rights granted to Curis under the collaboration agreement. The shares issued to Aurigene are subject to a lock-up agreement until January 18, 2017, with a portion of the shares being released from the lock-up in four equal bi-annual installments between now and that date.

The agreement provides that the parties will collaborate exclusively in immuno-oncology for an initial period of approximately two years, with the option for Curis to extend the broad immuno-oncology exclusivity.

In addition Curis has agreed to make payments to Aurigene as follows:

  • for the first two programs: up to $52.5 million per program, including $42.5 million per program for approval and commercial milestones, plus specified approval milestone payments for additional indications, if any;
  • for the third and fourth programs: up to $50 million per program, including $42.5 million per program for  approval and commercial milestones, plus specified approval milestone payments for additional indications, if any; and
  • for any program thereafter: up to $140.5 million per program, including $87.5 million per program in approval and commercial milestones, plus specified approval milestone payments for additional indications, if any.

Curis has agreed to pay Aurigene royalties on any net sales ranging from high single digits to 10% in territories where it successfully commercializes products and will also share in amounts that it receives from sublicensees depending upon the stage of development of the respective molecule.

About IRAK4:

Interleukin-1 receptor-associated kinase 4, or IRAK4 is a signaling kinase that becomes inappropriately activated in certain cancers including activated B cell-diffuse large B cell lymphoma (ABC-DLBCL), an aggressive form of lymphoma with poor prognosis. There appears to be a mechanistic link with IRAK4 in ABC-DLBCL where these tumors from approximately 35% of patients harbor oncogenic mutations in the MYD88 gene, which encodes an adaptor protein that interacts directly with IRAK4. MYD88 mutations appear to constitutively activate the IRAK4 kinase complex, driving pro-survival pathways in ABC-DLBCL disease. Oncogenic MYD88 mutations have also been identified in other cancers, including in over 90% of patients with Waldenström’s Macroglobulinemia as well as in a subset of patients with chronic lymphocytic leukemia (CLL).

About Curis, Inc.

Curis is a biotechnology company focused on the development and commercialization of novel drug candidates for the treatment of human cancers. Curis’ pipeline of drug candidates includes CUDC-907, a dual HDAC and PI3K inhibitor, CUDC-427, a small molecule antagonist of IAP proteins, and Debio 0932, an oral HSP90 inhibitor. Curis is also engaged in a collaboration with Genentech, a member of the Roche Group, under which Genentech and Roche are developing and commercializing Erivedge®, the first and only FDA-approved medicine for the treatment of advanced basal cell carcinoma. For more information, visit Curis’ website at www.curis.com.

About Aurigene

Aurigene is a specialized, discovery stage biotechnology company, developing novel and best-in-class therapies to treat cancer and inflammatory diseases. Aurigene’s Programmed Death pathway program is the first of several immune checkpoint programs that are at different stages of discovery and preclinical development. Aurigene has partnered with several large- and mid-pharma companies in the United States and Europe and has delivered multiple clinical compounds through these partnerships. With over 500 scientists, Aurigene has collaborated with 6 of the top 10 pharma companies. Aurigene is an independent, wholly owned subsidiary of Dr. Reddy’s Laboratories Ltd. (NYSE:RDY). For more information, please visit Aurigene’s website at http://aurigene.com/.

Small Molecule IRAK4 Kinase Inhibitor)

Innate immune responses mediated through Toll-like receptors or certain interleukin receptors are important mediators of the body’s initial defense against foreign antigens, while their dysregulation is associated with certain inflammatory conditions.  Toll-like receptor and interleukin receptor signaling through the adaptor protein MYD88, results in the assembly and activation of IRAK4, initiating a signaling cascade that induces cytokine and survival factor expression mediated by the transcription factor NFκB. More recently, components of this pathway are recognized to be genetically altered and have important roles in specific human cancers.  Toll-like receptor and interleukin receptor signaling through the adaptor protein MYD88, results in the assembly and activation of IRAK4, initiating a signaling cascade that induces cytokine and survival factor expression mediated by the transcription factor NFκB.  MYD88 gene mutations are shown to occur in approximately 30% of Activated B-Cell (ABC) subtype of diffuse large B-cell lymphomas (DLBCL)1,2 and in over 90% of the B-cell malignancy Waldenstrom’s macroglobulinemia.3  Due to IRAK4’s central role in these signaling pathways, it is considered an attractive target for generation of therapeutics to treat these B-cell malignancies as well as certain inflammatory diseases.

As part of the collaboration with Aurigene, in October 2015 we exercised our option to exclusively license a program of orally-available, small molecule inhibitors of IRAK4 kinase, including the development candidate, CA-4948.  Curis expects to file an IND and initiate clinical testing of CA-4948 in patients with advanced hematologic cancers during the second half of 2016.

1Nature. 2011; 470(7332):115–1192Immunology and Cell Biology. 2011; 89(6):659–6603N Engl J Med. 30, 2012; 367(9):826–833

CLIP

In November 2015, preclinical data were presented at the 2015 AACR-NCI-EORTC Molecular Targets and Cancer Therapeutics Conference in Boston, MA

Aurigene Collaboration (IRAK4 Inhibitor):

In October 2015, Curis exercised its option to exclusively license a program of orally available small molecule inhibitors of IRAK4 kinase, a serine/threonine kinase involved in innate immune responses as well as in certain hematologic cancers. The Company has since designated the development candidate as CA-4948 and expects to file an IND application for this molecule during 2016.

In November 2015, Curis’ collaborator Aurigene presented preclinical data from the IRAK4 program at the 2015 AACR-NCI-EORTC Molecular Targets and Cancer Therapeutics Conference in Boston, MA. This presentation included data from chemically distinct series of small molecule compounds with potent IRAK4 inhibitory activity in biochemical assays as well as in in vivo preclinical models, including MYD88 mutant DLBCL xenograft tumor models as well as a model of inflammatory disease.

CLIP

In April 2014, preclinical data presented at the CHI’s Ninth Drug Discovery Chemistry Conference in San Diego, CA, showed the compounds in vivo to have activity down to 10 mg/kg .

CLIP

10:50 Novel IRAK4 Inhibitors for Oncology and Inflammation
Susanta SamajdarSusanta Samajdar, Ph.D., Research Director, Medicinal Chemistry, Aurigene Discovery Technologies Limited
This presentation will discuss the discovery and optimization of hit series, some preliminary in vivo data, combination therapy strategy, present focus and further advancements.
CLIP

April 24-25 2014
Drug Discovery Chemistry – CHI’s Ninth Annual Conference: Fifth Annual Kinase inhibitor Chemistry, San Diego, CA, USA

Novel IRAK4 inhibitors

Susanta Samajdar from Aurigene Discovery Technologies presented the discovery of new IRAK4 (IL-1 receptor-associated kinase 4) inhibitors. Research began with a HTS campaign using two types of libraries: rationally designed novel scaffolds by hopping and morphing of known IRAK4 inhibitors and novel scaffolds identified by virtual screening of drug-like commercial library. A benzoxazol series was identified and crystallography was used to help their design. Lead optimization culminated in the identification of very potent compounds (AU-2807 and AU-2202) in cell assay (inflammation pathway and oncology pathway, respectively). The compounds were also active against Flt3 and KDR. Some PD in vivo data using LPS and TNFalpha release were presented in which the compound showed activity down to 10 mg/kg: no other in vivo model data were disclosed, but it was mentioned that studies in the CIA (collagen induced arthritis) model was ongoing. Dr Samajdar answered to three questions, one related to IRAK1 selectivity (the answer was that the compound is fully selective against IRAK1 and IRAK2). It was also mentioned that the compounds have a PBB higher than 98%. And the last question was related to the synergetic effect with BTK inhibitor in activated B-cell like diffuse large B-cell lymphoma, and this effect was observed with these compounds.

Susanta Samajdar

Research Director at Aurigene Discovery Technologies

PATENT

http://www.google.com/patents/WO2013042137A1?cl=en

Compound-6: Synthesis of 6′-amino-N-(5-(cyclopropyIamino)-2-morpholinobenzo [d]oxazoI-6-yl)-[2,3′-bipyridine]-6-carboxamide.

Step_l^N-cyclopropyl-2-morpholino-6-nitrobenzo[d]oxazol-5-amine.

N-cyclopropyl-2-moφholino-6-nitrobenzo[d]oxazol-5-amine(0.7g,70%) was prepared from 5-fluoro-2-mo holino-6-nitrobenzo[d]oxazole(lg,Intermediate-2) by treating with cyciopropanamine in sealed tube at 100°C for 8-14h. The progress of the reaction was monitored by TLC. After the reaction was completed, it was extracted with water (15ml) and dichioromethane (2x 15ml). The organic layer was collected, washed with brine, dried over sodium sulfate and concentrated under reduced pressure to get the crude. MS (ES) m/e 305(M+1, 50%).

Steg2:6-bromo-N-(5-(cyclopropylamino)-2-morpholinobenzo[d]oxazol-6-yl)

picolinamide.

Step Π and ii):The process of these steps are adopted from step 2 and step 3 of compound- 1.

Step3:6′-amino-N-(5-(cvclopropvlamino)-2-morpholinobenzord]oxazol-6-yl)-r2,3′- bipyridine]-6-carboxamide.

(i) N-(4-methoxybenzyl)-5-(4,4,5,5-tetramethyl-l ,3,2-dioxaborolan-2-yl)pyridin

Na2C03, Pd(dppf)Cl2, ACN, H20, 80-100°C, 8-14h; TFA, 60-70°C, 8-14h.

6′-amino-N-(5-(cyclopropylamino)-2-mo holinobenzo[d]oxazol-6-yl)-[2,3′-bipyridine]-6- carboxamide (0.03g,61%) was prepared from 6-bromo-N-(5-(cyclopropyIamino)-2- moφholinobenzo[d]o azoI-6-yl)picolinamide(0.07g, step-3) by following the same process used in step-1 and 2 of compound-3.

Ή NMR (400 MHz, DMSO-< ):6 1 1.63 (s, IH), 8.90 (s, IH), 8.61 (s, IH), 8.55 (s, IH), 8.37- 8.03 (m, 2H), 7.39 (s, IH), 6.80-6.62 (s, IH), 3.80-3.59 (m, 15H), 2.88-2.64 (m, 2H). MS (ESI): 472 (M+l , 60%).

PATENT

WO2015104688

Example 13

6′-amino-N-(2-morphol ne]-6-carboxamide

Step-1: Synthesis of 6-chloro thiazolo[4,5-c]pyridine-2(3H)-thione

Using the same reaction conditions as described in step 1 of example 1, 4,6-dichloropyridin-3-amine (1.3 g, 7 mmol) was cyclised using potassium ethyl xanthate (2.55 g, 15 mmol) in DMF (25mL) at 150°C for 8h to afford the title compound (1.3 g, 86.6 %) as a light brown solid.

1HNMR (400 MHz, DMSO-d6): δ 14.2-14.0 (b, 1H), 8.274 (s, 1H), 7.931 (s, 1H); LCMS: 100%, m/z = 201.3 (M+l)+.

Step-2: Synthesis of 4-(6-chloro thiazolo[4,5-c]pyridin-2-yl) morpholine

To a suspension of 6-chlorothiazolo[4,5-c]pyridine-2(3H)-thione (0.3 g, 1.16 mmol) in

DCM (4 mL), oxalyl chloride (0.2 mL, 2.38 mmol) and DMF (1.5 mL) were added at 0°C. The resulting mixture was slowly allowed to warm to room temperature and stirred there for 1 h. The reaction mixture was again cooled to 0°C and triethyl amine (0.66 mL, 4.76 mmol) and morpholine (0.13 mL, 1.75 mmol) were added. The reaction mixture was stirred at RT for 1 h and quenched with water and extracted with ethyl acetate. The combined organic layers were washed with water, brine, dried over sodium sulphate and concentrated under reduced pressure. The crude material was purified by column chromatography (EtOAc/n-hexanes 3:7) to afford the title compound (0.14 g, 39.6 %) as a light brown solid.

1H NMR (400 MHz, DMSO-d6): δ 8.47 (s, 1H), 8.04 (s, 1H), 3.74-3.72 (m, 4H), 3.61-3.59 (m, 4H); LCMS: m/z = 256.1 (M+l)+.

Step-3: Synthesis of 6′-amino-/V-(2-morpholino thiazolo [4,5-c]pyridin-6-yl)-[2,3′-bipyridine]-6-carboxamide

Using the same reaction conditions as described in step 4 of example 12, 4-(6-chlorothiazolo[4,5-c] pyridin-2-yl) morpholine (0.081 g, 0.32 mmol), was coupled with tert-butyl (6-carbamoyl-[2,3′-bipyridin]-6′-yl)carbamate (intermediate 2) (0.1 g, 0.32 mmol) using cesium carbonate (0.21 g, 0.64 mmol), XantPhos (0.028g, 0.047mmol) and Pd2(dba)3 (0.015 mg, 0.015 mmol) in toluene : dioxane (2:2mL) to get the crude product. The resultant crude was purified by 60-120 silica gel column chromatography using 2% methanol in DCM as eluent. Further the resultant crude was purified by prep HPLC to afford title compound (0.01 g, 6 %) as an off-white solid.

1H NMR (400 MHz, DMSO-d6): δ 10.65 (s, 1H), 8.88 (d, 1H), 8.85 (dd, 1H), 8.71 (s, 1H), 8.55 (s, 1H), 8.22-8.13 (m, 4 H), 7.09 (d, 1H), 3.73 (t, 4H), 3.58 (t, 4H). LCMS: 100%, m/z = 434.2 (M+l)+.

Example 11

(S)-2-(2-methylpyridin-4-yl)-N-(2-morpholino-5-(pyrrolidin-3-ylamino)oxazolo[4,5-b]pyridin-6-yl)oxazole-4-carboxamide

Step l:Preparation of (S)-tert-butyl 3-((2-morpholino-6-nitrooxazolo[4,5-b]pyridin-5-yl)amino)pyrrolidine- 1 -carboxylate

A solution of 5-chloro-2-morpholino-6-nitrooxazolo[4,5-b]pyridine (300mg, 1.0563 mmol) (S)-tert-butyl 3 -aminopyrrolidine- 1 -carboxylate (237mg, 1.267 mmol) and potassium carbonate (292mg, 2.112 mmol) in DMF (2mL) was heated at 100°C for 2h. Reaction was quenched with ice water and filtered the solid. The resultant crude was purified by 60-120 silica gel column chromatography using 1 % methanol in DCM as eluent to obtain the title compound (350mg, 76.25%). LCMS: m/z: 435.4 (M+l)+.

Step 2:Preparation of (S)-tert-butyl 3-((6-amino-2-morpholinooxazolo[4,5-b]pyridin-5-yl)amino)pyrrolidine- 1 -carboxylate

Using the same reaction conditions as described in step 5 of example 1, (S)-tert-butyl 3- ((2-morpholino-6-nitrooxazolo[4,5-b]pyridin-5-yl)amino)pyrrolidine-l -carboxylate (350mg, 0.806 mmol) was reduced with zinc dust (422mg, 6.451 mmol) and ammonium chloride (691mg, 12.903 mmol) in THF/methanol/H20 (10mL/2mL/lmL) to get the title compound (240mg, 71.8%). LCMS: m/z: 405.2 (M+l)+.

Step 3:Preparation of (S)-tert-butyl 3-((6-(2-(2-methylpyridin-4-yl)oxazole-4-carboxamido)-2-morpholinooxazolo[4,5-b]pyridin-5-yl)amino)pyrrolidine-l-carboxylate

Using the same reaction conditions as described in step 6 of example 1, (S)-tert-butyl 3-((6-amino-2-morpholinooxazolo[4,5-b]pyridin-5-yl)amino)pyrrolidine-l -carboxylate (115mg, 0.284 mmol), was coupled with 2-(2-methylpyridin-4-yl)oxazole-4-carboxylic acid (70mg, 0.341 mmol) using EDCI.HCl (82mg, 0.426 mmol), HOBt (58mg, 0.426 mmol), DIPEA (0.199mL, 1.138 mmol) in DMF (2mL) to afford the title compound (lOOmg, 59.52%). LCMS: m/z: 591.4 (M+l)+.

Step 4: Preparation of (S)-2-(2-methylpyridin-4-yl)-N-(2-morpholino-5-(pyrrolidin-3-ylamino)oxazolo[4,5-b]pyridin-6-yl)oxazole-4-carboxamide

Using the same reaction conditions as described in step 8 of example 1, (S)-tert-butyl 3- ((6-(2-(2-methylpyridin-4-yl)oxazole-4-carboxamido)-2-morpholinooxazolo[4,5-b]pyridin-5-yl)amino)pyrrolidine-l -carboxylate (lOOmg, 0.169 mmol) was deprotected using methanolic HC1 (5mL) to get the crude product. This was then purified by prep HPLC to get the title compound (9mg, 10.84%).

1HNMR (CDCI3, 400MHz): δ 9.91 (s, 1H), 8.78 (s, 1H), 8.74-8.73 (d, 1H), 8.45 (s, 1H), 7.82 (s, 1H), 7.76-7.74 (d, 1H), 4.50 (s, 1H), 4.04-4.03 (d, 4H), 3.30-3.00 (m, 7H), 2.70 (s, 3H), 2.40-1.80 (m, 4H), 1.00-0.08 (m, 1H). LCMS: 100%, m/z = 491.3 (M+l)+.

REFERENCES

http://www.curis.com/images/stories/pdfs/posters/Aurigene_IRAK4_AACR-NCI-EORTC_2015.pdf

http://www.curis.com/images/stories/pdfs/posters/Aurigene_IRAK4_AACR_20150421.pdf

1Nature. 2011; 470(7332):115–119

2Immunology and Cell Biology. 2011; 89(6):659–660

3N Engl J Med. 30, 2012; 367(9):826–833

April 2014, preclinical data presented at the CHI’s Ninth Drug Discovery Chemistry Conference in San Diego, CA

November 2015, preclinical data were presented at the 2015 AACR-NCI-EORTC Molecular Targets and Cancer Therapeutics Conference in Boston, MA

http://pubs.acs.org/doi/abs/10.1021/jm5016044

http://cancerres.aacrjournals.org/content/75/15_Supplement/3646

2015 Apr 18-22; Philadelphia, PA. Philadelphia (PA): AACR; Cancer Res 2015;75(15 Suppl):Abstract nr 3646. doi:10.1158/1538-7445.AM2015-3646

////////IRAK4 Kinase Inhibitor, Curis,  Aurigene,  CA 4948, AU 4948, CA-4948, AU-4948, 1428335-77-6

c21ccc(cc1sc(n2)N3CCOCC3)NC(c4nc(ccc4)c5ccc(nc5)N)=O


सुकून उतना ही देना प्रभू, जितने से जिंदगी चल जाये। औकात बस इतनी देना, कि औरों का भला हो जाये।
DRUG APPROVALS BY DR ANTHONY MELVIN CRASTO …..FOR BLOG HOME CLICK HERE

Join me on Linkedin

View Anthony Melvin Crasto Ph.D's profile on LinkedIn

Join me on Facebook FACEBOOK

Join me on twitterFollow amcrasto on Twitter
Join me on google plus Googleplus
Join me on Researchgate

Anthony Melvin Crasto Dr.

 amcrasto@gmail.com


09b37-misc2b027LIONEL MY SON
He was only in first standard in school when I was hit by a deadly one in a million spine stroke called acute transverse mylitis, it made me 90% paralysed and bound to a wheel chair, Now I keep him as my source of inspiration and helping millions, thanks to millions of my readers who keep me going and help me to keep my son happy

सुकून उतना ही देना प्रभू, जितने से
जिंदगी चल जाये।
औकात बस इतनी देना,
कि औरों का भला हो जाये।
Read all about Organic Spectroscopy on ORGANIC SPECTROSCOPY INTERNATIONAL  

Curis and Aurigene’s AUPM 170, CA 170


1,2,4-oxadiazole and 1 ,2,4-thiadiazole compounds of formula (I):

ONE EXAMPLE

STR3

EXAMPLES

STR3

PREDICTED AUPM 170, CA 170, AUPM-170, CA-170

STR3

Synthesis coming………….

WATCH THIS SPACE

Aurigene Discovery Technologies Limited INNOVATOR

Curis with the option to exclusively license Aurigene’s orally-available small molecule antagonist of programmed death ligand-1 (PD-L1) in the immuno-oncology field

Addressing immune checkpoint pathways is a well validated strategy to treat human cancers and the ability to target PD-1/PD-L1 and other immune checkpoints with orally available small molecule drugs has the potential to be a distinct and major advancement for patients.

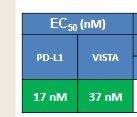
Through its collaboration with Aurigene, Curis is now engaged in the discovery and development of the first ever orally bioavailable, small molecule antagonists that target immune checkpoint receptor-ligand interactions, including PD-1/PD-L1 interactions.  In the first half of 2016, Curis expects to file an IND application with the U.S. FDA to initiate clinical testing of CA-170, the first small molecule immune checkpoint antagonist targeting PD-L1 and VISTA.  The multi-year collaboration with Aurigene is focused on generation of small molecule antagonists targeting additional checkpoint receptor-ligand interactions and Curis expects to advance additional drug candidates for clinical testing in the coming years. The next immuno-oncology program in the collaboration is currently targeting the immune checkpoints PD-L1 and TIM3.

In November 2015, preclinical data were reported. Data demonstrated tha the drug rescued and sustained activation of T cells functions in culture. CA-170 resulted in anti-tumor activity in multiple syngeneic tumor models including melanoma and colon cancer. Similar data were presented at the 2015 AACR-NCI-EORTC Molecular Targets and Cancer Therapeutics Conference in Boston, MA

By August 2015, preclinical data had been reported. Preliminary data demonstrated that in in vitro studies, small molecule PD-L1 antagonists induced effective T cell proliferation and IFN-gamma production by T cells that were specifically suppressed by PD-L1 in culture. The compounds were found to have effects similar to anti-PD1 antibodies in in vivo tumor models

(Oral Small Molecule PD-L1/VISTAAntagonist)

Certain human cancers express a ligand on their cell surface referred to as Programmed-death Ligand 1, or PD-L1, which binds to its cognate receptor, Programmed-death 1, or PD-1, present on the surface of the immune system’s T cells.  Cell surface interactions between tumor cells and T cells through PD-L1/PD-1 molecules result in T cell inactivation and hence the inability of the body to mount an effective immune response against the tumor.  It has been previously shown that modulation of the PD-1 mediated inhibition of T cells by either anti-PD1 antibodies or anti-PD-L1 antibodies can lead to activation of T cells that result in the observed anti-tumor effects in the tumor tissues.  Therapeutic monoclonal antibodies targeting the PD-1/PD-L1 interactions have now been approved by the U.S. FDA for the treatment of certain cancers, and multiple therapeutic monoclonal antibodies targeting PD-1 or PD-L1 are currently in development.

In addition to PD-1/PD-L1 immune regulators, there are several other checkpoint molecules that are involved in the modulation of immune responses to tumor cells1.  One such regulator is V-domain Ig suppressor of T-cell activation or VISTA that shares structural homology with PD-L1 and is also a potent suppressor of T cell functions.  However, the expression of VISTA is different from that of PD-L1, and appears to be limited to the hematopoietic compartment in tissues such as spleen, lymph nodes and blood as well as in myeloid hematopoietic cells within the tumor microenvironment.  Recent animal studies have demonstrated that combined targeting/ blockade of PD-1/PD-L1 interactions and VISTA result in improved anti-tumor responses in certain tumor models, highlighting their distinct and non-redundant functions in regulating the immune response to tumors2.

As part of the collaboration with Aurigene, in October 2015 Curis licensed a first-in-class oral, small molecule antagonist designated as CA-170 that selectively targets PD-L1 and VISTA, both of which function as negative checkpoint regulators of immune activation.  CA-170 was selected from the broad PD-1 pathway antagonist program that the companies have been engaged in since the collaboration was established in January 2015.  Preclinical data demonstrate that CA-170 can induce effective proliferation and IFN-γ (Interferon-gamma) production (a cytokine that is produced by activated T cells and is a marker of T cell activation) by T cells that are specifically suppressed by PD-L1 or VISTA in culture.  In addition, CA-170 also appears to have anti-tumor effects similar to anti-PD-1 or anti-VISTA antibodies in multiple in vivo tumor models and appears to have a good in vivo safety profile.  Curis expects to file an IND and initiate clinical testing of CA-170 in patients with advanced tumors during the first half of 2016.

Curis, Inc.

Jan 21, 2015

Curis and Aurigene Announce Collaboration, License and Option Agreement to Discover, Develop and Commercialize Small Molecule Antagonists for Immuno-Oncology and Precision Oncology Targets

— Agreement Provides Curis with Option to Exclusively License Aurigene’s Antagonists for Immuno-Oncology, Including an Antagonist of PD-L1 and Selected Precision Oncology Targets, Including an IRAK4 Kinase Inhibitor —

— Investigational New Drug (IND) Application Filings for Both Initial Collaboration Programs Expected this Year —

— Curis to issue 17.1M shares of its Common Stock as Up-front Consideration —

— Management to Host Conference Call Today at 8:00 a.m. EST —

LEXINGTON, Mass. and BANGALORE, India, Jan. 21, 2015 (GLOBE NEWSWIRE) — Curis, Inc. (Nasdaq:CRIS), a biotechnology company focused on the development and commercialization of innovative drug candidates for the treatment of human cancers, and Aurigene Discovery Technologies Limited, a specialized, discovery stage biotechnology company developing novel therapies to treat cancer and inflammatory diseases, today announced that they have entered into an exclusive collaboration agreement focused on immuno-oncology and selected precision oncology targets. The collaboration provides for inclusion of multiple programs, with Curis having the option to exclusively license compounds once a development candidate is nominated within each respective program. The partnership draws from each company’s respective areas of expertise, with Aurigene having the responsibility for conducting all discovery and preclinical activities, including IND-enabling studies and providing Phase 1 clinical trial supply, and Curis having responsibility for all clinical development, regulatory and commercialization efforts worldwide, excluding India and Russia, for each program for which it exercises an option to obtain a license.

The first two programs under the collaboration are an orally-available small molecule antagonist of programmed death ligand-1 (PD-L1) in the immuno-oncology field and an orally-available small molecule inhibitor of Interleukin-1 receptor-associated kinase 4 (IRAK4) in the precision oncology field. Curis expects to exercise its option to obtain exclusive licenses to both programs and file IND applications for a development candidate from each in 2015.

“We are thrilled to partner with Aurigene in seeking to discover, develop and commercialize small molecule drug candidates generated from Aurigene’s novel technology and we believe that this collaboration represents a true transformation for Curis that positions the company for continued growth in the development and eventual commercialization of cancer drugs,” said Ali Fattaey, Ph.D., President and Chief Executive Officer of Curis. “The multi-year nature of our collaboration means that the parties have the potential to generate a steady pipeline of novel drug candidates in the coming years. Addressing immune checkpoint pathways is now a well validated strategy to treat human cancers and the ability to target PD-1/PD-L1 and other immune checkpoints with orally available small molecule drugs has the potential to be a distinct and major advancement for patients. Recent studies have also shown that alterations of the MYD88 gene lead to dysregulation of its downstream target IRAK4 in a number of hematologic malignancies, including Waldenström’s Macroglobulinemia and a subset of diffuse large B-cell lymphomas, making IRAK4 an attractive target for the treatment of these cancers. We look forward to advancing these programs into clinical development later this year.”

Dr. Fattaey continued, “Aurigene has a long and well-established track record of generating targeted small molecule drug candidates with bio-pharmaceutical collaborators and we have significantly expanded our drug development capabilities as we advance our proprietary drug candidates in currently ongoing clinical studies. We believe that we are well-positioned to advance compounds from this collaboration into clinical development.”

CSN Murthy, Chief Executive Officer of Aurigene, said, “We are excited to enter into this exclusive collaboration with Curis under which we intend to discover and develop a number of drug candidates from our chemistry innovations in the most exciting fields of cancer therapy. This unique collaboration is an opportunity for Aurigene to participate in advancing our discoveries into clinical development and beyond, and mutually align interests as provided for in our agreement.  Our scientists at Aurigene have established a novel strategy to address immune checkpoint targets using small molecule chemical approaches, and have discovered a number of candidates that modulate these checkpoint pathways, including PD-1/PD-L1. We have established a large panel of preclinical tumor models in immunocompetent mice and can show significant in vivo anti-tumor activity using our small molecule PD-L1 antagonists.  We are also in the late stages of selecting a candidate that is a potent and selective inhibitor of the IRAK4 kinase, demonstrating excellent in vivo activity in preclinical tumor models.”

In connection with the transaction, Curis has issued to Aurigene approximately 17.1 million shares of its common stock, or 19.9% of its outstanding common stock immediately prior to the transaction, in partial consideration for the rights granted to Curis under the collaboration agreement. The shares issued to Aurigene are subject to a lock-up agreement until January 18, 2017, with a portion of the shares being released from the lock-up in four equal bi-annual installments between now and that date.

The agreement provides that the parties will collaborate exclusively in immuno-oncology for an initial period of approximately two years, with the option for Curis to extend the broad immuno-oncology exclusivity.

In addition Curis has agreed to make payments to Aurigene as follows:

  • for the first two programs: up to $52.5 million per program, including $42.5 million per program for approval and commercial milestones, plus specified approval milestone payments for additional indications, if any;
  • for the third and fourth programs: up to $50 million per program, including $42.5 million per program for  approval and commercial milestones, plus specified approval milestone payments for additional indications, if any; and
  • for any program thereafter: up to $140.5 million per program, including $87.5 million per program in approval and commercial milestones, plus specified approval milestone payments for additional indications, if any.

Curis has agreed to pay Aurigene royalties on any net sales ranging from high single digits to 10% in territories where it successfully commercializes products and will also share in amounts that it receives from sublicensees depending upon the stage of development of the respective molecule.
About Immune Checkpoint  Modulation and Programmed Death 1 Pathway

Modulation of immune checkpoint pathways has emerged as a highly promising therapeutic approach in a wide range of human cancers. Immune checkpoints are critical for the maintenance of self-tolerance as well as for the protection of tissues from excessive immune response generated during infections. However, cancer cells have the ability to modulate certain immune checkpoint pathways as a mechanism to evade the immune system. Certain immune checkpoint receptors or ligands are expressed by various cancer cells, targeting of which may be an effective strategy for generating anti-tumor activity. Some immune-checkpoint modulators, such as programmed death 1 (PD-1) protein, specifically regulate immune cell effector functions within tissues. One of the mechanisms by which tumor cells block anti-tumor immune responses in the tumor microenvironment is by upregulating ligands for PD-1, such as PD-L1. Hence, targeting of PD-1 and/or PD-L1 has been shown to lead to the generation of effective anti-tumor responses.
About Curis, Inc.

Curis is a biotechnology company focused on the development and commercialization of novel drug candidates for the treatment of human cancers. Curis’ pipeline of drug candidates includes CUDC-907, a dual HDAC and PI3K inhibitor, CUDC-427, a small molecule antagonist of IAP proteins, and Debio 0932, an oral HSP90 inhibitor. Curis is also engaged in a collaboration with Genentech, a member of the Roche Group, under which Genentech and Roche are developing and commercializing Erivedge®, the first and only FDA-approved medicine for the treatment of advanced basal cell carcinoma. For more information, visit Curis’ website at www.curis.com.

About Aurigene

Aurigene is a specialized, discovery stage biotechnology company, developing novel and best-in-class therapies to treat cancer and inflammatory diseases. Aurigene’s Programmed Death pathway program is the first of several immune checkpoint programs that are at different stages of discovery and preclinical development. Aurigene has partnered with several large- and mid-pharma companies in the United States and Europe and has delivered multiple clinical compounds through these partnerships. With over 500 scientists, Aurigene has collaborated with 6 of the top 10 pharma companies. Aurigene is an independent, wholly owned subsidiary of Dr. Reddy’s Laboratories Ltd. (NYSE:RDY). For more information, please visit Aurigene’s website at http://aurigene.com/.

POSTER

STR3

STR3

STR3

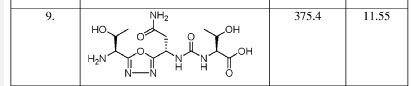
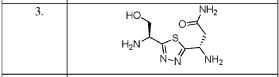
WO2011161699, WO2012/168944, WO2013144704 and WO2013132317 report peptides or peptidomimetic compounds which are capable of suppressing and/or inhibiting the programmed cell death 1 (PD1) signaling pathway.

PATENT

WO 2015033299

Example 5: Synthesis of

The compound was synthesised using similar procedure as depicted in Example 4 (compound 4) using D-amino acids are linked up in reverse order. Boc-D-Thr(‘Bu)-OH was used in place of Boc-Ser(‘Bu)-OH, Fmoc-D-Asn(trt)-OH in place of Fmoc-Asn(trt)-OH and H-D-Ser(‘Bu)-0’Bu was used in place of H-Thr^Bu^O’Bu to yield 0.3 g crude material of the title compound. The cmde solid material was purified using preparative HPLC described under experimental conditions. LCMS: 361.3 (M+H)+. HPLC: tR = 13.58 min.

Example 8: Synthesis of

The compound was synthesised using similar procedure as depicted in Example 2 (compound 2) using Fmoc-Glu(0’Bu)-OH instead of Fmoc-Asn(Trt)-OH to get 0.4 g crude material of the title compound. The crude solid material was purified using preparative HPLC described under experimental conditions. LCMS: 362.1 (M+H)+. HPLC: tR = 13.27 min.

PATENT

WO2015033301

Example 3: Synthesis of compound 3

Step 3a:

3a

Lawesson’s reagent (2.85 g, 7.03 mmol) was added to a solution of compound 2e (4 g, 4.68 mmol) in THF (40 mL) and stirred at 75°C for 4 h. The completeness of the reaction was confirmed by TLC analysis. The reaction mixture was evaporated under reduced

pressure and the obtained residue was partitioned between ice water and ethyl acetate. The organic layer was washed with NaHCC>3 solution followed brine solution. The organic layer was dried over Na2S04, filtered and evaporated under reduced pressure to get residue which was further purified by silica gel column chromatography (eluent: 0-5% ethyl acetate in hexane) to afford 2.7 g of compound 3a (Yield: 67.66%). LCMS: 852.3 (M+H)+,

Step 3

3a 3b

Fmoc group on compound 3a was deprotected by adding diethylamine (3.8 mL) to the solution of compound 3a (1 g, 1.17 mmol) in CH2CI2 (3.8 mL). The reaction mixture was stirred at room temperature for 30 min. The resulting solution was concentrated in vacuum to get a thick gummy residue. The crude compound was purified by neutral alumina column chromatography (eluent: 0-50% ethyl acetate in hexane then 0-5% methanol in chloroform) to attain 0.62 g of compound 3b. LCMS: 630.5 (M+H)+.

Step 3c

To a solution of compound 3b (0.6 g) in CH2CI2 (7.5 mL), trifluoroacetic acid (2.5 mL) and catalytic amount of triisopropylsilane were added and stirred at room temperature for 3 h. The resulting solution was concentrated in vacuum to get 0.13 g of compound 3 which was purified by preparative HPLC method described under experimental conditions. LCMS: 232.3 (M+H)+.

Example 1: Synthesis of compound 1

Step la:

Potassium carbonate (7.9 g, 57.39 mmol) and Methyl iodide (1.3 mL, 21.04 mmol) were added to a solution of compound la (5.0 g, 19.13 mmol) in DMF (35 mL) and stirred at room temperature for 2 h. The completeness of the reaction was confirmed by TLC analysis. The reaction mixture was partitioned between water and ethyl acetate. Organic layer was washed with water, brine, dried over Na2S04 and evaporated under reduced pressure to get 5.0 g of compound lb (Yield: 96.1%). LCMS: 176.1 (M-Boc)+.

Step lb:

Hydrazine hydrate (7.2 mL) was added to a solution of compound lb (5.0 g, 18.16 mmol) in methanol (30 mL) and stirred at room temperature for 2 h. The completeness of the reaction was confirmed by TLC analysis. The reaction mixture was evaporated under reduced pressure, the residue obtained was partitioned between water and ethyl acetate. Organic layer was washed with water, brine, dried over Na2S04 and evaporated under reduced pressure to get 4.0 g of compound lc (Yield: 80.0%). LCMS: 276.3 (M+H)+. Step lc:

NMM (0.67 ml, 6.52 mmol) was slowly added to a stirred solution of lc (1.2 g, 4.35 mmol), Id (1.43 g, 4.35 mmol), HOBt (0.7 g, 5.22 mmol) and EDC.HC1 (0.99 g, 5.22 mmol) in DMF (15 mL) at 0°C. The reaction mixture was stirred at room temperature for 12 h. The completeness of the reaction was confirmed by TLC analysis. The reaction was quenched with ice and the solid precipitated was filtered and dried under vacuum to obtain 2.0 g of pure product le (Yield: 83.3%). LCMS: 591.5 (M+Na)+.

St

1 e

1f

To a stirred solution of le (1.5 g, 2.63 mmol) in dry THF (15.0 mL) and DMF (5.0 mL) triphenylphosphine (1.38 g, 5.27 mmol) and iodine (1.33 g, 5.27 mmol) were added at 0°C. After the iodine was completely dissolved, Et3N (1.52 mL, 10.54 mmol) was added to this reaction mixture at ice cold temperature. Reaction mixture was allowed to attain room temperature and stirred for 4 h. The completeness of the reaction was confirmed by TLC analysis. The reaction was quenched with ice water and extracted with ethyl acetate. Organic layer was washed with saturated sodium thiosulphate and brine solution.

The separated Organic layer was dried over Na2SC>4 and evaporated under reduced pressure to get residue, which was further purified by silica gel column chromatography (eluent: 30% ethyl acetate in hexane) to afford 0.8 g of compound If (Yield: 55%). LCMS: 551.3 (M+H)+.

Step le:

1f i g

Fmoc group was deprotected by the addition of diethylamine (20.0 mL) to a solution of compound If (0.8 g, 1.45 mmol) in CH2CI2 (20.0 mL) at 0°C. The reaction was stirred at room temperature for 2 h. The resulting solution was concentrated in vacuum to get a thick gummy residue. The crude compound was purified by neutral alumina column chromatography (eluent: 2% methanol in chloroform) to afford 0.38 g of compound lg (Yield: 80.0%): LCMS: 329.4 (M+H)+.

Step If:

ig 1 i

Compound lg (0.38 g, 1.16 mmol), TEA (0.33 mL, 2.32 mmol) dissolved in DMF (10 mL) were added drop wise to a solution of lh (0.55 g, 1.39 mmol) at 0°C for urea bond formation and the mixture was stirred at room temperature for 2 h. The completeness of the reaction was confirmed by TLC analysis. The reaction was quenched with ice water, the solid precipitated was filtered and dried under vacuum to get crude compound, which was further purified by silica gel column chromatography (eluent: 0-35% ethyl acetate in hexane) to get 0.4 g of product li (Yield: 59.7%). LCMS: 586.4 (M+H)+.

Step lg:

BocHN’ IJ, H LT Y~™

1

To a solution of compound li (0.4 g, 0.68 mmol) in CH2CI2 (5 m L), trifluoro acetic acid (5 mL) and catalytic amount of triisopropylsilane were added and stirred at room temperature for 3 h to remove the acid sensitive protecting groups. The resulting solution was concentrated under nitrogen and the solid material was purified by preparative HPLC method as described under experimental conditions (Yield: 0.05 g). LCMS: 318.0 (M+H)+; HPLC: tR= 10.96 min.

Synthesis of compound lh (N02-C6H4-OCO-Thr(tBu)- 0¾u):

To a solution of 4-nitrophenylchloroformate (4.79 g, 23.77 mmol) in DCM (25.0 mL) was added a solution of H-Thr(tBu)-OtBu (5.0 g, 21.61 mmol) TEA (6.2 mL, 43.22 mmol) in CH2CI2 (25 mL) slowly at 0°C and allowed to stir for 30 min. The completion of the reaction was confirmed by TLC analysis. After completion of reaction it was diluted with DCM and washed with 1.0 M of citric acid followed by 1.0 M sodium carbonate solution. The organic layer was dried over Na2S04 and evaporated under reduced pressure to afford crude compound 1 h, which was further purified by silica gel column chromatography (eluent: 0-5% ethyl acetate in hexane) to get 3.0 g of product lh. jH NMR (CDCI3, 400 MHz): £1.17 (s, 9H), 1 .28 (d, 3H), .50 (s, 9H), 4.11 (m, 1 H), 4.28 (m, 1H , 5.89 (d, 1H), 7.37 (d, 2H), 8.26 (d, 2H).

Pottayil Sasikumar

Murali Ramachandra

Brahma Reddy V, Thomas Antony, Murali Ramachandra, Venkateshwar Rao G, Wesley Roy Balasubramanian, Kishore Narayanan, Samiulla DS, Aravind AB, and Shekar Chelur.

REFERENCES

US20150073024

WO2011161699A2 27 Jun 2011 29 Dec 2011 Aurigene Discovery Technologies Limited Immunosuppression modulating compounds
WO2012168944A1 21 Dec 2011 13 Dec 2012 Aurigene Discovery Technologies Limited Therapeutic compounds for immunomodulation
WO2013132317A1 4 Mar 2013 12 Sep 2013 Aurigene Discovery Technologies Limited Peptidomimetic compounds as immunomodulators
WO2013144704A1 28 Mar 2013 3 Oct 2013 Aurigene Discovery Technologies Limited Immunomodulating cyclic compounds from the bc loop of human pd1

http://www.curis.com/pipeline/immuno-oncology/pd-l1-antagonist

http://www.curis.com/images/stories/pdfs/posters/Aurigene_PD-L1_VISTA_AACR-NCI-EORTC_2015.pdf

////////Curis and Aurigene,  AUPM 170, CA 170, AUPM-170, CA-170, PD-L1, VISTA antagonist

ND 2158


(2S)-2-hydroxy-3-[(3R)-12-{[(1r,4r)-4-(morpholin-4-yl)cyclohexyl]oxy}-7-thia-9,11-diazatricyclo[6.4.0.0²,⁶]dodeca-1(12),2(6),8,10-tetraen-3-yl]propanamide

S)-2-hydroxy-3-((R)-4-(((lr,4R)-4-morpholinocyclohexyl)oxy)-6,7-dihydro-5H-cyclopenta [4,5] thieno [2,3-d] pyrimidin-5-yl)propanamide

 CAS 1388896-07-8
C22 H30 N4 O4 S
5H-​Cyclopenta[4,​5]​thieno[2,​3-​d]​pyrimidine-​5-​propanamide, 6,​7-​dihydro-​α-​hydroxy-​4-​[[trans-​4-​(4-​morpholinyl)​cyclohexyl]​oxy]​-​, (αS,​5R)​-
Molecular Weight446.56

STR3

ND 2158

IRAK4, 446.2

C22H30N4O4S

Company Nimbus Therapeutics LLC
Description IL-1 receptor-associated kinase 4 (IRAK4) inhibitor
Molecular Target Interleukin-1 receptor-associated kinase 4 (IRAK4)
Mechanism of Action Interleukin-1 receptor-associated kinase 4 (IRAK4) inhibitor
Therapeutic Modality Small molecule

ND-2158 is a potent and selective experimental inhibitor of IRAK4 described in patent WO2013106535 [2] and in a poster presented at the American College of Rheumatology meeting in 2012 (Abstract #1062 in Supplement: Abstracts of the American College of Rheumatology & Association of Rheumatology Health Professionals, Annual Scientific Meeting, November 9-4, 2012 Washington DC, Volume 64, Issue S10, Page S1-S1216).

PATENT

WO2013106535

http://www.google.com/patents/WO2013106535A1?cl=en

Figure imgf000085_0001

Figure imgf000086_0001

Scheme II

Example 88: (S)-l-((R)-4-(((lr,4R)-4-morpholinocyclohexyl)oxy)-6,7-dihydro- 5H-cyclopenta[4,5]thieno[2,3-d]pyrimidin-5-yl)butan-2-ol (1-64) and Example 89: (R)-l- ((R)-4-(((lr,4R)-4-morpholinocyclohexyl)oxy)-6,7-dihydro-5H-cyclopenta[4,5]thieno[2,3-

Synthesis of compound 88.1. Note: For the preparation of the starting material compound 29.2, please see Example 29. A solution of

yl)cyclohexyl]oxy]-7-thia-9,l l-diazatricyclo[6.4.0.0[2,6]]dodeca-l(8),2(6),9,l l-tetraen-3- yl]ethan-l-ol (190 mg, 0.47 mmol, 1.00 equiv) in 10 mL of dichloromethane was added Dess- Martin periodinane at 0 °C in a water/ice bath under nitrogen. The resulting mixture was stirred for 2 h at room temperature. After completion of the reaction, the mixture was then diluted with saturated aqueous sodium bicarbonate and extracted with 3 x 30 mL of ethyl acetate. The combined organic layers were dried over sodium sulfate and concentrated under vacuum. The residue was applied onto a silica gel column with ethyl acetate/petroleum ether (1 :5 to 1 : 1) to afford 2-[(3Λ)-12-[[4-^ο 1ιο1ϊη-4-γ1)ογο1ο1ιβχγ1]οχγ]-7-ωΕ-9,11- diazatricyclo[6.4.0.0[2,6]]dodeca-l(8),2(6),9,l l-tetraen-3-yl]acetaldehyde (130 mg, 69%) as a colorless oil. MS (ES): m/z 402 [M+H]+.

Synthesis of Compound 1-64 and Compound 1-65. A solution of [(3i?)-12-[[4- (moφholin-4-yl)cyclohexyl]oxy]-7-thia-9,l l-diazatricyclo[6.4.0.0[2,6]]dodeca-l(8),2(6),9,l l- tetraen-3-yl]acetaldehyde (130 mg, 0.32 mmol, 1.00 equiv) in 5 mL of anhydrous THF was added bromo(ethyl)magnesium (1 M in THF, 0.62 mL, 2.0 equiv) dropwise at 0 °C under nitrogen. The resulting solution was stirred for 4 h at room temperature and then quenched by the addition of saturated aqueous NH4CI and extracted with 3 x 50 mL of DCM/i-PrOH (3:1). The combined organic layers was dried over anhydrous sodium sulfate and concentrated under vacuum. The crude product (150 mg) was purified by preparative HPLC under the following conditions (SHIMADZU): column: SunFire Prep C18, 19*150 mm 5um; mobile phase: water with 0.05% NH4CO3 and CH3CN (6.0% CH3CN up to 54.0% in 25 min); UV detection at 254/220 nm to afford (S)-l-((R)-4-(((lr,4R)-4-moφholinocyclohexyl)oxy)-6,7-dihydro-5H- cyclopenta[4,5]thieno[2,3-d]pyrimidin-5-yl)butan-2-ol (11.8 mg) and (R)-l-((R)-4-(((lr,4R)-4- mo holinocyclohexyl)oxy)-6,7-dihydro-5H-cyclopenta[4,5]thieno[2,3-d]pyrimidin-5-yl)butan- 2-ol (23.9 mg) as white solids.

Example 88 (1-64): MS: 432 (M+H)+. ¾ NMR (300 MHz, CDC13) S 8.47 (s, 2H), 5.24-5.20 (m, 1H), 3.75-3.58 (m, 5H), 3.06-2.93 (m, 2H), 2.70-2.61 (m, 4H), 2.28-1.98 (m, 3H), 1.59-1.41 (m, 10H), 1.28-1.23 (m, 2H),0.95-0.85 (m, 3H).

Example 89 (1-65): MS: 432 (M+H)+. ¾ NMR (300 MHz, CDC13) S 8.47 (s, 2H), 5.25 (m, 1H), 3.71-3.39 (m, 6H), 3.04-2.90 (m, 2H), 2.67-2.55 (m, 5H), 2.34-2.22 (m, 4H), 2.01- 1.81 (m, 3H), 1.64-1.39 (m, 7H), 0.94-0.92 (m, 3H).

WATCH OUT SYNTHESIS COMING…………

PATENT

WO 2014011906

https://www.google.co.in/patents/WO2014011906A2?cl=en

PATENT

WO-2014194242

https://www.google.com/patents/WO2014194242A2?cl=en

Example 49: Synthesis of Intermediate 49.1.

Image loading...

step 1 step 2

35.1 49.1 49.2 Image loading...

step 3 49 3

] Intermediate 49.3 was prepared from 35.1 in a manner analogous to the synthesis of 36.3. Isolated 150 mg of a white solid in 57% overall yield. MS (ES): m/z 402 [M+H]+.

Example 50: Synthesis of Intermediate 50.4.

Image loading...

49.3 50.1 50.2

Image loading...

50.3 50.4

Intermediate 50.4 was prepared from 49.3 in a manner analogous to the synthesis of 1-25, except that HCl/MeOH rather than TBAF/THF was used in the second step. Isolated 124 mg of a white solid in 48% overall yield. MS (ES): m/z 447 [M+H]+. 1H NMR (400 MHz, CDCls): δ 8.46 (s, 1H), 5.28-5.25 (m, 1H), 4.17-4.06 (m, 51H), 3.74-3.72 (m, 5H), 3.37-2.98 (m, 2H), 2.72-2.28 (m, 10H), 2.11-2.08 (m, 2H), 1.79-1.46 (m, 5H).

Example 51: Synthesis of (S)-2-hydroxy-3-((R)-4-(((lr,4R)-4- morpholinocyclohexyl)oxy)-6,7-dihydro-5H-cyclopenta [4,5] thieno [2,3-d] pyrimidin-5- yl)propanamide (1-34) and Example 52: Synthesis of (R)-2-hydroxy-3-((R)-4-(((lr,4R)-4- morpholinocyclohexyl)oxy)-6,7-dihydro-5H-cyclopenta [4,5] thieno [2,3-d] pyrimidin-5- yl)propanamide (1-44)

Image loading...

The racemic 50.4 (1.6 g, 96.5% purity) was separated by Chiral-HPLC with the following conditions (Gilson G x 281): column: Chiralpak AD-H, 2*25 cm Chiral-P(AD-H); mobile phase: phase A: hex (O. P/oDEA) (HPLC grade), phase B: IPA (HPLC grade), gradient: 30% B in 9 min; flow rate: 20 mL/min; UV detection at 220/254 nm. The former fractions (tR = 4.75 min) were collected and evaporated under reduced pressure and lyophilized overnight to afford 1-44 (520 mg) with 100% ee as a white solid. And the latter fractions (tR = 5.82 min) were handled as the former fractions to give the desired 1-34 (510 mg) with 99.6%> ee as a white solid. The ee values of the two isomers were determined by the chiral-HPLC with the following conditions (SHIMADZU-SPD-20A): column: Chiralpak AD-H, 0.46*25 cm, 5um (DAICEL); mobile phase: hex (0.1% TEA): IPA = 85:15; UV detection at 254 nm. Flow rate: 1.0 mL/min. tR (1-44) = 7.939 min and tR (1-34) = 11.918 min.

[00431] Analytical data for 1-44: MS: (ES, m/z) 447 [M+H]+. 1H NMR (400 MHz, CD3OD+CDCI3): δ 8.47 (s, 1H), 5.32-5.22 (m, 1H), 4.08 (dd, 1H), 4.89-4.62 (m, 5H), 3.20-3.10 (m, 1H), 3.05-2.95 (m, 1H), 2.75-2.55 (m, 5H), 2.44-2.38 (m, 2H), 2.34-2.28 (m, 3H), 2.10 (d, 2H), 1.82-1.62 (m, 3H), 1.58-1.40 (m, 2H).

Analytical data for 1-34: MS: (ES, m/z) 447 [M+H]+. 1H NMR (400 MHz, CDC13): δ 8.46 (s, 1H), 5.32-5.22 (m, 1H), 4.15 (t, 1H), 3.73 (t, 4H), 3.59 (td, 1H), 3.19-3.08 (m, 1H), 3.02- 2.92 (m, 1H), 2.78-2.70 (m, 1H), 2.69-2.60 (m, 4H), 2.58-2.20 (m, 5H), 2.10 (d, 2H), 1.75-1.63 (m, 3H), 1.53-1.40 (m, 2H).

Paper

http://pubs.acs.org/doi/abs/10.1021/jm5016044

Recent Advances in the Discovery of Small Molecule Inhibitors of Interleukin-1 Receptor-Associated Kinase 4 (IRAK4) as a Therapeutic Target for Inflammation and Oncology Disorders

Miniperspective

Nimbus Discovery, 25 First Street, Suite 404, Cambridge, Massachusetts 02141, United States
Schrödinger Inc., 120 West Forty-Fifth Street, New York, New York 10036, United States
J. Med. Chem., 2015, 58 (1), pp 96–110
DOI: 10.1021/jm5016044
Abstract Image

IRAK4, a serine/threonine kinase, plays a key role in both inflammation and oncology diseases. Herein, we summarize the compelling biology surrounding the IRAK4 signaling node in disease, review key structural features of IRAK4 including selectivity challenges, and describe efforts to discover clinically viable IRAK4 inhibitors. Finally, a view of knowledge gained and remaining challenges is provided.

STR3

  1. 78 Romero, D. L.; Robinson, S.; Wessel, M. D.; Greenwood, J. R. IRAK Inhibitors and Uses Thereof. WO201401902, January 16, 2014.

  2. 79.

    Harriman, G. C.; Romero, D. L.; Masse, C. E.; Robinson, S.; Wessel, M. D.; Greenwood, J. R. IRAK Inhibitors and Uses Thereof. WO2014011911A2, January 16, 2014.

  3. 80.

    Harriman, G. C.; Wester, R. T.; Romero, D. L.; Masse, C. E.; Robinson, R.; Greenwood, J. R. IRAK Inhibitors and Uses Thereof. WO2014011906A2, January 16, 2014
Patent ID Date Patent Title
US2013231328 2013-09-05 IRAK INHIBITORS AND USES THEREOF

PATENT

STR3

WO 2014194242

WO 2013106535

WO 2012097013

US20070155777 * Feb 21, 2007 Jul 5, 2007 Amgen, Inc. Antiinflammation agents
US20100041676 * Feb 18, 2010 Hirst Gavin C Kinase inhibitors
US20100143341 * Jun 21, 2006 Jun 10, 2010 Develogen Aktiengesellschaft Thienopyrimidines for pharmaceutical compositions
US20120015962 * Jan 19, 2012 Nidhi Arora PYRAZOLO[1,5a]PYRIMIDINE DERIVATIVES AS IRAK4 MODULATORS
US20120283238 * Nov 8, 2012 Nimbus Iris, Inc. Irak inhibitors and uses thereof
References
1. Chaudhary D, Robinson S, Romero DL. (2015)
Recent Advances in the Discovery of Small Molecule Inhibitors of Interleukin-1 Receptor-Associated Kinase 4 (IRAK4) as a Therapeutic Target for Inflammation and Oncology Disorders.
J. Med. Chem.58 (1): 96-110. [PMID:25479567]
2. Harriman GC, Wester RT, Romero DL, Robinson S, Shelley M, Wessel MD, Greenwood JR, Masse CE, Kapeller-Libermann R. (2013)
Irak inhibitors and uses thereof.
Patent number: WO2013106535. Assignee: Nimbus Iris, Inc.. Priority date: 18/07/2013. Publication date: 10/01/2012.

http://nimbustx.com/sites/default/files/uploads/posters/irak4_nimbus_acr_poster_2012_small.pdf

///////ND 2158, IRAK4, ND-2158, NIMBUS, 1388896-07-8

NC(=O)C(CC1CCc2c1c1c(ncnc1s2)OC1CCC(CC1)N1CCOCC1)O

C1CC(CCC1N2CCOCC2)OC3=C4C5=C(CCC5CC(C(=O)N)O)SC4=NC=N3

ND 2110


STR3

ND -2110

Molecular Formula: C21H28N4O3S
Molecular Weight: 416.53702 g/mol

2-[(3R)-12-{[(1r,4r)-4-(morpholin-4-yl)cyclohexyl]oxy}-7-thia-9,11-diazatricyclo[6.4.0.0²,⁶]dodeca-1(12),2(6),8,10-tetraen-3-yl]acetamide

1388894-17-4

C21 H28 N4 O3 S, 5H-​Cyclopenta[4,​5]​thieno[2,​3-​d]​pyrimidine-​5-​acetamide, 6,​7-​dihydro-​4-​[[trans-​4-​(4-​morpholinyl)​cyclohexyl]​oxy]​-​, (5R)​-
Molecular Weight416.54

ND-2110 is a potent and selective experimental inhibitor of IRAK4 described in patent WO2013106535 [2] and in a poster presented at the American College of Rheumatology meeting in 2012 (Abstract #1062 in Supplement: Abstracts of the American College of Rheumatology & Association of Rheumatology Health Professionals, Annual Scientific Meeting, November 9-4, 2012 Washington DC, Volume 64, Issue S10, Page S1-S1216).

Company Nimbus Therapeutics LLC
Description IL-1 receptor-associated kinase 4 (IRAK4) inhibitor
Molecular Target Interleukin-1 receptor-associated kinase 4 (IRAK4)
Mechanism of Action Interleukin-1 receptor-associated kinase 4 (IRAK4) inhibitor
Therapeutic Modality Small molecule

PATENT

WO2013106535

http://www.google.com/patents/WO2013106535A1?cl=en

Example 29: Synthesis of 2-((R)-4-(((lr,4R)-4-morpholinocyclohexyl)oxy)-6,7-

29.3 1-67

Synthesis of compound 29.1. 4-(Morpholin-4-yl)cyclohexan-l-ol (commercially available; 218 mg, 1.2 mmol, 1.50 equiv) was treated with NaH (60% dispersion in mineral oil, 128 mg, 3.2 mmol, 4 equiv) in freshly distilled tetrahydrofuran (15 mL) for 30 min at 0 °C in a water/ice bath under nitrogen. Then a solution of intermediate 25.1 (289 mg, 0.8 mmol, 1.00 equiv) in 5 mL of THF was added via syringe and the resulting solution was allowed to stir for an additional 3 h at 60 °C in an oil bath. The reaction was then quenched with saturated aqueous NH4CI and extracted with 3 x 50 mL of ethyl acetate. The combined organic layers were washed with brine, dried (Na2S04) and concentrated under vacuum. The residue was applied onto a silica gel column with ethyl acetate/petroleum ether (1:5-1:2) and purified to afford compound 29.1 (260 mg, 63%) as a colorless oil.

Synthesis of compound 29.2. To a solution of 29.1 (260 mg, 0.5 mmol, 1.0 equiv) in 10 mL of DCM was added 0.5 mL of concentrated hydrochloric acid in an ice/water bath. The resulting solution was stirred for 2 h and concentrated in vacuo. The residue was neutralized with saturated aqueous Na2C03 and extracted with 3 x 50 mL of ethyl acetate. The organic layers were combined, washed with brine, dried (Na2S04) and concentrated under reduced pressure. The residue was purified by column chromatography on silica gel with DCM MeOH (15:1) to afford the desired alcohol 29.2 (185 mg, 91%) as a colorless oil. [00416] Synthesis of compound 29.3. Alcohol 29.2 (185 mg, 0.46 mmol, 1.00 equiv) was oxidized with dipyridinium dichromate (752 mg, 2.00 mmol, 4.36 equiv) in 50 mL of DMF for 24 h at room temperature. The resulting solution was diluted with water and extracted with 3 x 50 mL of mixed solutions of CHC¾/iso-PrOH. The organic layers were combined, dried (Na2S04) and concentrated under vacuum. The residue was applied onto a silica gel column with dichloromethane/methanol (5:1 to 1:1) and purified to afford 105 mg (55%) of acid 29.3 as a yellow oil.

[00417] Synthesis of Compound 1-67. A 50 mL round-bottom flask containing a solution of acid 29.3 (105 mg, 0.25 mmol, 1.00 equiv), NH4C1 (80 mg, 1.50 mmol, 6.00 equiv), EDCI (57 mg, 0.3 mmol, 1.2 equiv), 4-dimethylaminopyridine (37 mg, 0.3 mmol, 1.2 equiv) and HOBt (40 mg, 0.3 mmol, 1.2 equiv) in 5 mL of anhydrous DMF was stirred for 24 h at room temperature. The resulting solution was diluted with water and extracted with 4 x 50 mL of mixed solution of CHCl3:iso-PrOH. The combined organic layers were concentrated under vacuum. The crude product was purified by preparative HPLC (SHIMADZU) under the following conditions: column: SunFire Prep C18, 19*150mm 5um; mobile phase: water (0.05% NH4CO3) and CH3CN (6.0% CH3CN up to 50.0% in 25 min); UV detection at 254/220 nm. The product-containing fractions were collected and concentrated to give Compound 1-67 (22.5 mg) as a white solid. ¾ NMR (300 MHz, CD3OD) δ 8.43 (s, 1H), 5.27-5.20 (m, 1H), 3.80-3.70 (m, 5H), 3.29-3.27 (m, 1H), 3.12-2.90 (m, 2H), 2.73-2.67 (m, 5H), 2.49-2.42 (m, 1H), 2.32-2.19 (m, 4H), 2.10-2.06 (d, 2H), 1.67-1.46 (m, 4H). MS: m/z 417 (M+H)+.

PATENT

http://www.google.com/patents/WO2012097013A1

Example 29: Synthesis of 2-((R)-4-(((lr,4R)-4-morpholinocyclohexyl)oxy)-6,7- dihydro-5H-cyclopenta[4,5]thieno[2,3-d]pyrimidin-5-yl)acetamide.

Page 280 of 407

2009184-0008

33b

Synthesis of compound 31b. 4-(Morpholin-4-yl)cyclohexan-l-ol (commercially available; 218 mg, 1.2 mmol, 1.50 equiv) was treated with NaH NMR (60% dispersion in mineral oil, 128 mg, 3.2 mmol, 4 equiv) in freshly distilled tetrahydrofuran (15 mL) for 30 min at 0 °C in a water/ice bath under nitrogen. Then a solution of intermediate Hb (289 mg, 0.8 mmol, 1.00 equiv) in 5 mL of THF was added via syringe and the resulting solution was allowed to stir for an additional 3 h at 60 °C in an oil bath. The reaction was then quenched with saturated aqueous NH4CI and extracted with 3 x 50 mL of ethyl acetate. The combined organic layers were washed with brine, dried (Na2S04) and concentrated under vacuum. The residue was applied onto a silica gel column with ethyl acetate/petroleum ether (1 :5-1 :2) and purified to afford compound 31b (260 mg, 63%) as a colorless oil.

Synthesis of compound 32b. To a solution of 31b (260 mg, 0.5 mmol, 1.0 equiv) in 10 mL of DCM was added 0.5 mL of concentrated hydrochloric acid in an ice/water bath. The resulting solution was stirred for 2 h and concentrated in vacuo. The residue was neutralized with saturated aqueous Na2C( j and extracted with 3 x 50 mL of ethyl acetate. The organic layers were combined, washed with brine, dried (Na2S04) and concentrated under reduced pressure. The residue was purified by column chromatography on silica gel with DCM/MeOH NMR ( 15: 1 ) to afford the desired alcohol 32b ( 185 mg, 91 %) as a colorless oil.

Synthesis of compound 33b. Alcohol 32b (185 mg, 0.46 mmol, 1.00 equiv) was oxidized with dipyridinium dichromate (752 mg, 2.00 mmol, 4.36 equiv) in 50 mL of DMF for

Page 281 of 407

2009184-0008 24 h at room temperature. The resulting solution was diluted with water and extracted with 3 x 50 mL of mixed solutions of CHCU/iso-PrOH. The organic layers were combined, dried (Na2S04) and concentrated under vacuum. The residue was applied onto a silica gel column with dichloromethane/methanol (5: 1 to 1 : 1 ) and purified to afford 105 mg (55%) of acid 33b as a yellow oil.

Synthesis of Compound. A 50 mL round-bottom flask containing a solution of acid 33b (105 mg, 0.25 mmol, 1.00 equiv), NH4C1 (80 mg, 1.50 mmol, 6.00 equiv), EDCI (57 mg, 0.3 mmol, 1.2 equiv), 4-dimethylaminopyridine (37 mg, 0.3 mmol, 1.2 equiv) and HOBt (40 mg, 0.3 mmol, 1.2 equiv) in 5 mL of anhydrous DMF was stirred for 24 h at room temperature. The resulting solution was diluted with water and extracted with 4 x 50 mL of mixed solution of CHCI3: iso-PrOH. The combined organic layers were concentrated under vacuum. The crude product was purified by preparative HPLC (SHIMADZU) under the following conditions: column: SunFire Prep C I 8, 19* 150mm 5um; mobile phase: water (0.05% Ν¾∞3) and CH3CN (6.0% CH3CN up to 50.0% in 25 min); UV detection at 254/220 nm. The product containing fractions were collected and concentrated to give the product (22.5 mg) as a white solid. Ή MR (300 MHz, CD3OD) δ 8.43 (s, 1H), 5.27-5.20 (m, 1H), 3.80-3.70 (m, 5H), 3.29-3.27 (m, 1 H), 3.12-2.90 (m, 2H), 2.73-2.67 (m, 5H), 2.49-2.42 (m, 1H), 2.32-2.19 (m, 4H), 2.10-2.06 (d, 2H), 1.67- 1.46 (m, 4H). MS: m/z 417 (M+H)+.

Paper

http://pubs.acs.org/doi/abs/10.1021/jm5016044

Recent Advances in the Discovery of Small Molecule Inhibitors of Interleukin-1 Receptor-Associated Kinase 4 (IRAK4) as a Therapeutic Target for Inflammation and Oncology Disorders

Miniperspective

Nimbus Discovery, 25 First Street, Suite 404, Cambridge, Massachusetts 02141, United States
Schrödinger Inc., 120 West Forty-Fifth Street, New York, New York 10036, United States
J. Med. Chem., 2015, 58 (1), pp 96–110
DOI: 10.1021/jm5016044
Abstract Image

IRAK4, a serine/threonine kinase, plays a key role in both inflammation and oncology diseases. Herein, we summarize the compelling biology surrounding the IRAK4 signaling node in disease, review key structural features of IRAK4 including selectivity challenges, and describe efforts to discover clinically viable IRAK4 inhibitors. Finally, a view of knowledge gained and remaining challenges is provided.

 STR3
  1. 78 Romero, D. L.; Robinson, S.; Wessel, M. D.; Greenwood, J. R. IRAK Inhibitors and Uses Thereof. WO201401902, January 16, 2014.

  2. 79.

    Harriman, G. C.; Romero, D. L.; Masse, C. E.; Robinson, S.; Wessel, M. D.; Greenwood, J. R. IRAK Inhibitors and Uses Thereof. WO2014011911A2, January 16, 2014.

  3. 80.

    Harriman, G. C.; Wester, R. T.; Romero, D. L.; Masse, C. E.; Robinson, R.; Greenwood, J. R. IRAK Inhibitors and Uses Thereof. WO2014011906A2, January 16, 2014

STR3

WO 2014194245

WO 2014194201

WO 2014194242

WO 2013106535

WO 2012097013

1. Chaudhary D, Robinson S, Romero DL. (2015)
Recent Advances in the Discovery of Small Molecule Inhibitors of Interleukin-1 Receptor-Associated Kinase 4 (IRAK4) as a Therapeutic Target for Inflammation and Oncology Disorders.
J. Med. Chem.58 (1): 96-110.
2. Harriman GC, Wester RT, Romero DL, Robinson S, Shelley M, Wessel MD, Greenwood JR, Masse CE, Kapeller-Libermann R. (2013)
Irak inhibitors and uses thereof.
Patent number: WO2013106535C1CC(CCC1N2CCOCC2)OC3=C4C5=C(CCC5CC(=O)N)SC4=NC=N3. Assignee: Nimbus Iris, Inc.. Priority date: 18/07/2013. Publication date: 10/01/2012.

http://nimbustx.com/sites/default/files/uploads/posters/irak4_nimbus_acr_poster_2012_small.pdf

///////ND-2110, ND 2110. IRAK4, NIMBUS, GTPL8802

NC(=O)CC1CCc2c1c1c(ncnc1s2)OC1CCC(CC1)N1CCOCC1

C1CC(CCC1N2CCOCC2)OC3=C4C5=C(CCC5CC(=O)N)SC4=NC=N3

CN-128 for the treatment of thelassemia and iron overload


Figure imgf000011_0002

STR3

CN-128

(R)-3-Hydroxy-1-(1-hydroxy-3-benzyl propyl-2-)2-methyl pyridine-4(1H)-one

IND Filing

CN-128 is potentially for the treatment of thelassemia and iron overload.

Zhejiang University, 浙江大学

CAS No. 1335282-04-6

C15 H17 N O3, 4(1H)​-​Pyridinone, 3-​hydroxy-​1-​[(1R)​-​1-​(hydroxymethyl)​-​2-​phenylethyl]​-​2-​methyl-
Molecular Weight, 259.30

Many diseases in humans and animals are caused by excessive accumulated metals, such as iron. Among such diseases, excess iron is accumulated in various tissues, which is called iron overload disorders, formerly known as siderosis Haemorrhagic. Excess iron has the following sources: 1) long-term blood transfusion; 2) the gastrointestinal system absorbing excess iron, because stimulated by diseases such as anemia. It is necessary to repeat transfusion for some patients with severe anemia, for example, β-thalassemia, as well as other anemia requiring transfusion therapy. Excessive iron absorption from the gastrointestinal tract usually occurs in hemochromatosis patients and in anemia patients who do not require blood transfusion, such as thalassemia intermedia. If iron overload disease is not treated, it will result in severe tissue damage, especially the liver, heart and endocrine organs, and ultimately lead to death. Iron chelators can remove and clear excess iron from such organs, relieve symptoms and reduce the corresponding mortality.

Desferrioxamine (DFO) is an effective iron chelator for a long time. However, in the treatment of the diseases mentioned above, the biggest disadvantage regarding DFO and its salts is its poor oral absorption capability. So, administration is achieved with a slow injection method (8∼12h/day), patients need to wear a portable drug delivery device during treatment, such as mounting the syringe on a mechanical pressing device. This method is inconvenient, and also expensive, which largely limits the utilization of DFO, especially for thalassemia-prone areas, such as Mediterranean, Middle East, and India &South East Asia, it plays no role in treatment of malaria in world-wide and sickle cell anemia in some African countries, which is a very serious problem to the populations there. Image loading...

UK Patent No. 2, 13, 807, US Patent No. 4, 585, 780 and other scientific research have reported the treatment of iron overload symptoms by using 3-hydroxypyridin-4-one derivatives, especially in some pathological symptoms, such as thalassemia, sickle cell anemia, aplastic anemia in children, and idiopathic hemochromatosis, usually, treatment of the first three diseases includes frequent regular blood transfusion. 3-Hydroxypyridin-4-one derivatives, especially CP20 (commercial named Ferriprox) is employed to treat systemic iron overload disorders, and also to treat certain diseases associated with local iron overload distribution, although such patients do not show symptoms of systemic iron overload, i.e. inhibition free radical mediated reactions caused by excess iron ions in certain neurodegenerative diseases and cancer diseases. A serious limitation of CP20 is that the hydroxyl group at 3′ position is vulnerable to glycosylation, which reduces the half-life of this compound (approximately 2∼3 h). So it requires a high dosage, which is associated with obvious side effects. Image loading...

EP0120669 discloses compounds with a 3-hydroxypyrid-4-one in which the H attached to the N atom is substituted by an aliphatic acyl group, or an aliphatic hydrocarbon group, these groups can be further substituted, but not by aromatic groups and their use against illnesses related to iron overload. Molenda et al. disclose in Journal of Medicinal Chemistry 1994, 37, pages 4363-4370 chiral 3-hydroxy-pyridin-4-one compound 6 as enhancing iron excretion.

US Patent No. 6, 465, 604 described a series of 3,5-diphenyl-1,2,4-triazole compounds, wherein including Exjade (commercial name), which has strong affinity to Fe(III), However, its active groups contain two negatively charged oxygen ions and a carboxyl group; it is a tridentate ligand while chelating Fe(III), which forms a Fe-L2 type complex, possessing three unit negative charges itself, that is bad for their discharge from cells/tissues. Moreover, one of the active groups is a nitrogen atom with a lone pair of electrons, Exjade may have a negative effect on the balance of Zn(II) in vivo, at the same time because it has two phenolic hydroxyl groups in different positions (forming intramolecular hydrogen bonds structure similar to cis/trans isomerization), it can be complexed to several zinc ions to form high molecular weight polymers complexes, which is not conducive to its discharge from the cells either. Image loading...

Absorption, distribution, metabolism and excretion of chiral medicines are largely related to the 3D structures of their chiral centers. For drug absorption, chiral compounds entering cells via active transport mechanism are usually carried by special transport proteins, their recognition of enantiomers can be different, resulting in different absorption of enantiomers. For drug distribution, the binding effects of plasma protein and tissues are also somewhat stereoselective, leading to different in vivo distribution of enantiomers; for stereoselective of drug metabolism refers to when the substrate is biotransformated, the pathway and speed of enantiomer metabolism by biological systems can be different. One enantiomer may show ascendant metabolism, and therefore it is of great significance to the indicators including drug transformation and in vivo half-life. Glomerular filtration, tubular secretion and reabsorption of chiral drugs to clear the chiral drugs, having stereoselectivity, while the glomerular filtration rate is closely related to drug’s selectivity to binding plasma protein, so discharge style of enantiomers (urine / feces percentage) and the rate is also different.

Therefore, the qualitative difference of the interactions of a pair of enantiomers with various binding sites may exist or not, and the quantitive difference may exist (strong or weak), which results in the different activities between enantiomers. Thus the selection of optical enantiomers for medical use, requires a comprehensive study of metabolic activity, toxicology and pharmacokinetic properties etc. Thus the chiral nature of the 3-hydroxypyridin-4-one derivatives described in this patent has an important role on in vivo iron chelation.

The effectiveness of many oral 3-hydroxy-4-one derivatives drugs are subject to metabolic reaction of the 3-hydroxy moiety, which may be quickly glycosylated (see Reaction I). The hydroxypyridone after glycosylation loses the ability to chelate Fe(III). We can effectively inhibit glycosylation reaction by introducing hydroxyl groups to alkyl substituted residues on the pyridine ring. In addition, the partition coefficient of 3-hydroxy-pyridin-4-one derivatives has a great impact on the in vivo distribution and toxic effects. We have introduced various alkyl groups to the chiral point of the compound, in order to modify their lipophilicity, i.e. a phenyl group connected to the chiral point in compound IV-b while in IV-a it is a methyl group, and thus compound IV-b is relatively more lipophilic, and easier to penetrate through cell membranes of various tissues and critical barriers such as the blood-brain barrier and the placental barrier, thus affecting its in vivo distribution. Thus increase of hydroxyl groups can affect the intestinal absorption capacity, by introducing a large alkyl group, intestinal absorption of 3-hydroxy-pyridin-4-one derivatives can be enhanced. Image loading...

Reaction I

Image loading...

scheme is as follows: Image loading... Image loading...

Example7. (R)-3-Hydroxy-1-(1-hydroxy-3-benzyl propyl-2-)2-methyl pyridine-4(1H)-one, Number: CN128.

Image loading...

60 g 3-phenyloxy-2-methyl-4H-pyran-one(Example 1) was dissolved in 150 mL n-butanol, then 83.7 g D-phenylalaninol was added in. After thoroughly mixing, the solution was refluxed at 118°C for 36 h. After cooling and filtration, products were purified by silica gel column chromatography with Eluent ethanol: acetic ester=1:40. After Elution, light brown solid was obtained after rotary evaporation, which was then dissolved into 150 mL ethanol and 15 mL water, then it was hydrogenated and debenzylated with 5% Pd/C as catalyst, the solvent was removed under rotary evaporation, the remaining solid was recrystallized with methanol and ether, leading to 25.25 g light yellow solid. The yield was 35.1%. The free alkali’s 1HNMR (DMSO-d6): δ 2.00 (s, 3H), 2.98 (dd, J1=14, J2=5.5, 1H), 3.11 (dd, J1=14, J2=5, 1H), 3.73 (m, 2H), 4.54 (m, 1H), 6.21 (d, J=7, 1H), 7.17 (m, 5H), 7.87 (d, J=7.5, 1H).

PATENT

CN 102190644

http://www.google.com/patents/CN102190644B?cl=en

Zhejiang University

\\\\\\\\\\\\CN-128 , ind filed,  thelassemia,  iron overload, zhejiang
c1ccc(cc1)CC(N\2/C=C\C(/C(=C/2C)O)=O)CO

Debiopharm and Aurigene dual c-src / jak inhibitors


SCHEMBL2237115.png

STR3str4

Debio 1142

Jak2 tyrosine kinase inhibitor; Src tyrosine kinase inhibitor

N-[4-methyl-3-[2-[4-(4-methylpiperazin-1-yl)anilino]-5-oxo-7,8-dihydropyrido[4,3-d]pyrimidin-6-yl]phenyl]-3-(trifluoromethyl)benzamide

Molecular Formula: C33H32F3N7O2
Molecular Weight: 615.64809 g/mol
1332328-01-4
Benzamide, N-​[3-​[7,​8-​dihydro-​2-​[[4-​(4-​methyl-​1-​piperazinyl)​phenyl]​amino]​-​5-​oxopyrido[4,​3-​d]​pyrimidin-​6(5H)​-​yl]​-​4-​methylphenyl]​-​3-​(trifluoromethyl)​-

Debiopharm S.A., Aurigene Discovery Technologies Ltd.

ALLISTER Andrès MC, Maximilien Murone,Saumitra Sengupta, Shankar Jayaram Shetty

https://www.google.co.in/patents/WO2011101806A1?cl=en

Bicyclic compounds and their uses as dual c-src / jak inhibitors

STR3

Apr. 14 /PR Newswire/ –Debiopharm and Aurigene Sign Agreement for the Development and Commercialisation of Debio 1142, a Novel Inhibitor of an Undisclosed Oncology Pathway

LAUSANNE, Switzerland and BANGALORE, India, April 14, 2011 /PRNewswire/ — Debiopharm Group(TM) (Debiopharm), a global biopharmaceutical development specialist that focuses on serious medical conditions and particularly oncology, and Aurigene Discovery Technologies Ltd (Aurigene), a Bangalore-based drug discovery company, signed on March 23, 2011 an option and exclusive worldwide license agreement concerning the development and commercialisation of Debio 1142, a novel inhibitor of an undisclosed oncology pathway.

“We are very excited about this new collaboration with Aurigene. Their business model offers a one stop solution for structure guided drug design, lead optimisation and preclinical work. The Debio 1142 project aims at developing inhibitors targeting a key oncology pathway, which plays essential roles in various solid tumours, including resistance to chemotherapy” said Dr Rolland-Yves Mauvernay, president and founder of Debiopharm S.A.

“Coming as it does after a successful collaboration programme we already had with Debiopharm, and as a continuation of our close to 5 year association, the relationship between Debiopharm and Aurigene demonstrates the strategic fit between organisations with complimentary scientific skills. We are happy that we have the opportunity to continue to work with Debiopharm, in a unique business model that has been tailor-made to meet each partners’ needs” added CSN Murthy, CEO of Aurigene.

About Debiopharm Group

Debiopharm Group(TM) (Debiopharm) is a Swiss-based global biopharmaceutical group of companies with a focus on the development of prescription drugs that target unmet medical needs. The group in-licenses, develops and/or co-develops promising biological and small molecule drug candidates having reached clinical development phases I, II or III as well as earlier stage candidates. It develops its products for global registration and maximum commercial potential. The products are out-licensed to pharmaceutical partners for sales and marketing. Debiopharm Group is also active in the field of companion diagnostics with a view to progressing in the area of personalised medicine. Debiopharm independently funds the worldwide development of all of its products while providing expertise in pre-clinical and clinical trials, manufacturing, drug delivery and formulation, and regulatory affairs. For more information on Debiopharm Group(TM), please visit: http://www.debiopharm.com.

About Aurigene

Aurigene Discovery Technologies Limited is a Bangalore-based biotech focused on collaborative drug discovery with pharmaceutical and biotech companies on a risk-sharing basis. Aurigene has fully integrated drug discovery infrastructure, from Target to IND, along with strong in house structural biology and fragment based drug design capabilities. The company is engaged in over 20 discovery collaborations with US and European large and mid-pharma companies in Oncology, Inflammatory disorders and anti-infectives. For more information on Aurigene, please visit: http://www.aurigene.com.

PATENT

WO 2011101806

http://www.google.co.in/patents/WO2011101806A1?cl=en

PAPER

Journal of Chemical and Pharmaceutical Research (2014), 6(4), 1146-1152

http://jocpr.com/vol6-iss4-2014/JCPR-2014-6-4-1146-1152.pdf

REFERENCES

INDIAN PATENTS

7554/CHENP/2012

415/CHE/2010

https://www.debiopharm.com/our-business/pipeline.html

http://www.giiresearch.com/report/labd315710-debiopharm-international-sa-product-pipeline.html

Patent ID Date Patent Title
US2013143895 2013-06-06 BICYCLIC COMPOUNDS AND THEIR USES AS DUAL C-SRC / JAK INHIBITORS
US8440679 2013-05-14 Bicyclic compounds and their uses as dual c-SRC / JAK inhibitors

///////////Debio 1142, Jak2 tyrosine kinase inhibitor,  Src tyrosine kinase inhibitor, Debio-1142, Debiopharm S.A.Aurigene Discovery Technologies Ltd, 1332328-01-4

c21cnc(nc1CCN(C2=O)c3c(ccc(c3)NC(=O)c4cc(ccc4)C(F)(F)F)C)Nc5ccc(cc5)N6CCN(CC6)C

GDC-0084


GDC-0084
CAS#: 1382979-44-3
Chemical Formula: C18H22N8O2
Exact Mass: 382.1866

Synonym: RG7666; RG-7666; RG 7666; GDC-0084; GDC0084; GDC 0084.

IUPAC/Chemical Name: 5-(6,6-dimethyl-4-morpholino-8,9-dihydro-6H-[1,4]oxazino[4,3-e]purin-2-yl)pyrimidin-2-amine

Company Roche
Description Phosphoinositide 3-kinase (PI3K) inhibitor
Molecular Target Phosphoinositide 3-kinase (PI3K)
Mechanism of Action Phosphoinositide 3-kinase (PI3K) inhibitor
Therapeutic Modality Small molecule
Latest Stage of Development Phase I
Standard Indication Brain cancer
Indication Details Treat progressive or recurrent high-grade glioma
Regulatory Designation
Partner Genentech Inc.
  • Originator Genentech
  • Class Antineoplastics; Small molecules
  • Mechanism of Action 1 Phosphatidylinositol 3 kinase inhibitors
  • 28 Jan 2015 Discontinued – Phase-I for Glioma in Spain (unspecified route)
  • 28 Jan 2015 Discontinued – Phase-I for Glioma in USA (unspecified route)
  • 01 Jan 2015 Genentech completes a phase I trial in Glioma in USA and Spain (NCT01547546)

GDC-0084, also known as RG7666, is a phosphatidylinositol 3-kinase (PI3K) inhibitor with potential antineoplastic activity. PI3K inhibitor GDC-0084 specifically inhibits PI3K in the PI3K/AKT kinase (or protein kinase B) signaling pathway, thereby inhibiting the activation of the PI3K signaling pathway. This may result in the inhibition of both cell growth and survival in susceptible tumor cell populations. Activation of the PI3K signaling pathway is frequently associated with tumorigenesis.

str1

http://pubs.acs.org/doi/pdf/10.1021/acsmedchemlett.6b00005

Abstract Image

An improved, efficient process with a significantly reduced process mass intensity (PMI) led to the multikilogram synthesis of a brain penetrant PI3K inhibitor GDC-0084. Highlights of the synthesis include a phase transfer catalyzed annulation in water, an efficient Suzuki-Miyaura cross-coupling of a chloropyrimidine with an arylboronic acid using a low palladium catalyst loading, and the development of a controlled crystallization to provide the API. The process delivered GDC-0084 with low levels of both impurities and residual metals.

Development of an Efficient, Safe, and Environmentally Friendly Process for the Manufacture of GDC-0084

Small Molecule Process Chemistry, Small Molecule Analytical Chemistry, Genentech, Inc., A Member of the Roche Group, 1 DNA Way, South San Francisco, California 94080, United States
Org. Process Res. Dev., Article ASAP
DOI: 10.1021/acs.oprd.6b00011
Publication Date (Web): March 11, 2016
Copyright © 2016 American Chemical Society

//////GDC-0084

NC1=NC=C(C2=NC(N3CCOCC3)=C4N=C(C(C)(C)OCC5)N5C4=N2)C=N1

str1

str1

5-(6,6-Dimethyl-4-morpholino-8,9-dihydro-6H-[1,4]oxazino[4,3-e]purin-2-yl)pyrimidin-2-amine GDC-0084 

mp 211 °C; 1H NMR (500 MHz, DMSO-d6) δ 9.09 (s, 2H), 7.03 (s, 2H), 4.32–4.17 (m, 4H), 4.17–4.04 (m, 4H), 3.84–3.65 (m, 4H), 1.58 (s, 6H); 13C NMR (125 MHz, DMSO-d6) δ 163.8, 157.6, 154.2, 152.5, 151.3, 151.0, 120.3, 117.3, 73.7, 66.2, 57.8, 45.2, 41.5, 27.3. HRMS [M + H]+calcd for C18H22N8O2 383.1938; found 383.1945.

  1. The Discovery of Clinical Development Candidate GDC-0084, a Brain Penetrant Inhibitor of Class I Phosphoinositide 3-Kinases (PI3K) and mTOR.

    HeffronT.NdubakuC.SalphatiL.AlickeB.CheongJ.;DrobnickJ.EdgarK.GouldS.LeeL.LesnickJ.LewisC.NonomiyaJ.Pangj.PliseE.Sideris,S.WallinJ.WangL.ZhangX.OliveroA. ACS Med. Chem. Lett. 2016, , DOI: 10.1021/acsmedchemlett.6b00005

  2. 3.

    (a) Purine Derivatives Useful as PI3 Kinase Inhibitors. GoldsmithP.HancoxT. C.HudsonA.PeggN. A.KulagowskiJ. J.NadinA. J.PriceS. PCT Int. Appl. WO 2009053716 A1 Apr 30, 2009.

    (b) Preparation of Purine Derivatives with PI3K Inhibitory Activity and Methods of Use Thereof. CastanedoG.Chuckowree,I.FolkesA.SutherlinD. P.WanN. C. PCT Int. Appl. WO 2009146406 A1 Dec 3, 2009

The new Annex 16 “Certification by a Qualified Person and Batch Release” will become effective as of 15 April 2016.


DR ANTHONY MELVIN CRASTO Ph.D's avatarDRUG REGULATORY AFFAIRS INTERNATIONAL

The new Annex 16 is coming into Force

The new Annex 16 “Certification by a Qualified Person and Batch Release” will become effective as of 15 April 2016. The contents will reflect the coming state of expectations regarding the batch release.

see

http://www.gmp-compliance.org/enews_05188_The-new-Annex-16-is-coming-into-Force_15099,15432,Z-QAMPP_n.html

The new Annex 16 “Certification by a Qualified Person and Batch Release” will become effective as of 15 April 2016.

It is centrally pointed out that the main duty of a Qualified Person (QP) is the certification of batches. In this context, the QP must personally ensure that the responsibilities listed under Chapter 1.6 are fulfilled. Chapter 1.7 lists many other responsibilities to be guaranteed by the QP. However the related activities can be delegated and the QP can rely on the respective quality management systems. Yet, the “QP should have on-going assurance that this reliance is well founded” (1.7). The 21 responsibilites listed include amongst others:

  • The…

View original post 406 more words