Home » Uncategorized (Page 74)
Category Archives: Uncategorized
 DRUG APPROVALS BY DR ANTHONY MELVIN CRASTO
.....FOR BLOG HOME CLICK HERE
Flag and hits
ORGANIC SPECTROSCOPY

SUBSCRIBE

 Subscribe in a reader
Enter your email address:
Delivered by FeedBurner
Subscribe to New Drug Approvals by Email


Drafts of revised USP plastic packaging chapters and : removal of the biological reactivity test for oral and topical dosage forms


In a recent Pharmacopeial Forum two revised USP general chapters have been published for comment. With these drafts, the USP expert committee is removing the requirement for <87> Biological Reactivity Tests, In Vitro testing for packaging materials and systems for oral and topical dosage forms. Read more about the draft chapters of <661.1> Plastic Materials of Construction and <661.2> Plastic Packaging Systems for Pharmaceutical Use.testing for packaging materials and systems for oral and topical dosage forms. Read more about the draft chapters of <661.1> Plastic Materials of Construction and <661.2> Plastic Packaging Systems for Pharmaceutical Use.
read
In Pharmacopeial Forum 42(4) [Jun-Jul 2016] drafts of two revised USP general chapters <661.1> Plastic Materials of Construction and <661.2> Plastic Packaging Systems for Pharmaceutical Use have been published for comment. Deadline for comments is September 30, 2016. With these drafts, the USP General Chapters – Packaging and Distribution Expert Committee…
View original post 2,996 more words
EMA reviews Medicines manufactured at U.S. Company
Following the issuance of two Non-Compliance Reports for two sites of the US based company, EMA has started a review of medicines manufactured by Pharmaceutics International Inc., USA.
The European Medicines Agency (EMA) has started a review of medicines manufactured by Pharmaceutics International Inc., USA. This follows the issuance of two Non-Compliance Reports for two sites of the US based company after an inspection in February 2016 conducted by the MHRA (the medicines regulatory agency in the United Kingdom) which highlighted several shortcomings in relation to good manufacturing practice (GMP).
Pharmaceutics International Inc. manufactures the centrally authorised medicine Ammonaps (sodium phenylbutyrate) and is also the registered manufacturing site for some other medicines that have been authorised through national procedures in the European Union (EU).
This inspection which was a follow-up to an inspection in June 2015 aimed to assess whether corrective measures agreed previously had been appropriately implemented. It found…
View original post 201 more words
New bicalutamide/enzalutamide derivatives as antiproliferative agents for the treatment of prostate cancer
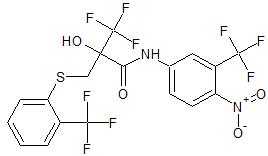
3,3,3-trifluoro-2-hydroxy-N-(4-nitro-3-(trifluoromethyl)phenyl)-2-(((2-(trifluoromethyl)phenyl)thio)methyl)propanamide
Cas 1929605-82-2

Dr Marcella Bassetto
Post Doctoral Research Associate
- bassettom@cardiff.ac.uk
- https://www.researchgate.net/profile/Marcella_Bassetto
- http://marcellabassetto.blogspot.in/
SYNTHESIS

Scheme .
Synthetic strategy used in the synthesis of 52. Reagents and conditions: (a) NaH (1 equiv.), THF, 0 °C to RT, 3 h; (b) KCN (1.2 equiv.), 25% H2SO4, 0 °C to RT, 20 h; c) HCl, AcOH, reflux, 24 h; (d) 8, SOCl2(1.3 equiv.), DMA, RT, 72 h.
3-Bromo-1,1,1-trifluoroacetone (48) was coupled with thiophenol 47 to afford 49, which was then converted into cyano derivative 50 using potassium cyanide and 25% sulfuric acid [16]. Intermediate 51 was obtained after refluxing 50 in concentrated HCl and glacial acetic acid. Coupling of 51 with commercially available 4-nitro-3-(trifluoromethyl)aniline 8yielded the desired amide 52.
Synthesis of 1,1,1-rifluoro-3-((2-(trifluoromethyl)phenyl)thio)propan-2-one (49)
To a mixture of NaH (10.47 mmol) in 10 mL anhydrous THF was added a solution of 2-(trifluoromethyl)benzenethiol (10.47 mmol) in 2mL anhydrous THF at 0 °C. This mixture was stirred for 20 min. 3-Bromo-1,1,1-trifluoropropan-2-one was then added dropwise to the mixture at 0 °C, the reaction was warmed to r.t. and stirred for 12 h. The mixture was filtered trough celite, the filtered pad was washed with THF, and the filtrate was evaporated to dryness. The residue was purified by flash column chromatography eluting with n-hexane/EtOAc 100:0 v/v increasing to n-hexane/EtOAc 85:15 v/v to give a pale yellow oil in 93% yield. 1H-NMR (CDCl3): d 7.76-7.69 (m, 2H), 7.60-7.53 (m, 1H), 7.42-7.38 (m, 1H), 3.44 (s, 2H). 19F-NMR (CDCl3): d -59.91 (s, 3F), -85.26 (s, 3F). 13C-NMR (CDCl3): d 189.6, 137.7, 135.9, 134.5, 133.2, 130.6, 129.6 (q, J= 26.3 Hz), 127.0 (q, J= 3.8 Hz), 124.3 (q, J= 4.1 Hz), 124.0 (q, J= 3.7 Hz), 94.4 (q, J= 30.4 Hz), 40.4.
Synthesis of 3,3,3-trifluoro-2-hydroxy-2-(((2-(trifluoromethyl)phenyl)thio)methyl)propanenitrile (50)
A 20% aqueous solution of H2SO4 (3.4 mL) was added dropwise to a mixture of 49 (11.03 mmol) and KCN (13.24 mmol) in 5 mL H2O at 0 °C. The reaction mixture was warmed to r.t. and stirred for 20 h. The mixture was then diluted with water (50 mL) and extracted with Et2O (3 x 150 mL). The organic extracts were washed with sat. aq. NaHCO3 and brine, dried over Na2SO4 and concentrated in vacuo. The residue was purified by flash column chromatography eluting with n-hexane/EtOAc 100:0 v/v increasing to n-hexane/EtOAc 95:5 v/v to give a pale yellow oil in 86% yield. 1H-NMR (CDCl3): d 7.80 (d, J= 7.8 Hz, 1H), 7.77-7.76 (m, 1H), 7.72-7.59 (m, 1H), 7.52-7.49 (m, 1H), 4.36 (bs, 1H), 3.58 (d, J= 14.6 Hz, 1H), 3.44 (d, J= 14.6 Hz, 1H). 19F-NMR (CDCl3): d -57.08 (s, 3F), -79.51 (s, 3F). 13C-NMR (CDCl3): d 135.4, 132.8, 132.5 (q, J= 30.1 Hz), 129.1, 128.7 (q, J= 5.5 Hz), 126.7, 124.9, 124.6, 122.6, 122.4, 120.4, 114.0, 71.4 (q, J= 32.9), 40.75.
1.1.1 Synthesis of 3,3,3-trifluoro-2-hydroxy-2-(((2-(trifluoromethyl)phenyl)thio)methyl)propanoic acid (51)
A mixture of 51 (6.89 mmol), concentrated HCl (23.4 mL) and AcOH (4.1 mL) was refluxed o.n. with vigorous stirring. The mixture was then diluted with water (100 mL) and extracted with Et2O (4 x 100 mL), which was in turn washed with sat. aq. NaHCO3 (4 x 100 mL). The water solution was acidified with concentrated HCl to pH 1 and extracted with Et2O (4x 150 mL). The Et2O extracts were dried over Na2SO4, filtered and concentrated to dryness to give a pale yellow waxy solid in 41% yield. 1H-NMR (CDCl3): d 9.57 (bs, 1H), 7.70 (d, J= 7.7 Hz, 1H), 7.67 (d, J= 7.7 Hz, 1H), 7.54-7.51 (m, 1H), 7.39-7.36 (m, 1H), 3.60 (s, 2H). 19F-NMR (CDCl3): d -60.10 (s, 3F), -77.7 (s, 3F). 13C-NMR (CDCl3): d 172.0, 134.1, 134.0, 131.2 (q, J= 30.1 Hz), 127.5, 126.7 (q, J= 5.6 Hz), 124.2 (q, J= 121.9 Hz), 121.9 (q, J= 126.7 Hz), 78.2 (q, J= 28.7 Hz), 37.7.
Synthesis of 3,3,3-trifluoro-2-hydroxy-N-(4-nitro-3-(trifluoromethyl)phenyl)-2-(((2-(trifluoromethyl)phenyl)thio)methyl)propanamide (52)
Thionyl chloride (1.16 mmol) was added dropwise to a stirring solution of 51 in anhydrous DMA at -10 °C under Ar atmosphere. The reaction mixture was stirred for 1 h, then a solution of 8 in 2 mL anhydrous DMA was added dropwise. The reaction mixture was warmed to r.t. and stirred for 72 h. The mixture was then diluted with sat. aq. NaHCO3 (40 mL) and extracted with Et2O (3 x 40 mL). The organic extracts were filtered trough celite, dried over Na2SO4 and evaporated to dryness. The residue was purified by flash column chromatography eluting with n-hexane/EtOAc 100:0 v/v increasing to n-hexane/EtOAc 80:20 v/v to give a pale yellow solid in 13% yield.
1H-NMR (CDCl3): d 8.93 (bs, 1H), 7.94 (d, J= 8.8 Hz, 1H), 7.87 (d, J= 2.2 Hz, 1H), 7.72 (d, J= 8.1 Hz, 1H), 7.69 (dd, J= 8.8 Hz, 2.2 Hz, 1H), 7.50-7.47 (m, 2H), 7.26-7.23 (m, 1H), 4.41 (s, 1H), 4.19 (d, 14.7 Hz, 1H), 3.45 (d, J= 14.7 Hz, 1H).
19F-NMR (CDCl3): d -59.7 (s, 3F), -60.12 (s, 3F), -77.4 (s, 3F).
13C-NMR (CDCl3): d 164.6, 143.8, 140.0, 134.7, 132.6, 131.1 (q, J= 29.8 Hz), 130.5, 128.3, 126.8 (q, J= 5.5 Hz), 126.7, 125.2 (q, J= 36.3 Hz), 124.5, 123.9, 122.6, 122.4, 122.2, 121.7, 120.4, 118.2 (q, J= 5.8 Hz), 76.3 (q, J= 27.8 Hz), 38.5.
MS [ESI, m/z]: 523.0 [M+H]+.
EI-HMRS (M-H)– found 521.0215, calculated for C18H0N2O4F9S 521.0218.
HPLC (method 1): retention time = 23.84 min.
clips
Prostate cancer (PC) is a leading cause of male death worldwide and it is the most frequently diagnosed cancer among men aged 65–74 [1]. The prognosis varies greatly, being highly dependent on a number of factors such as stage of diagnosis, race and age. Currently, PC treatment includes androgen deprivation, surgery, radiation, endocrine therapy and radical prostatectomy.
PC cell growth is strongly dependent on androgens, therefore blocking their effect can be beneficial to the patient’s health. Such outcomes can be achieved by antagonism of the androgen receptor (AR) using anti-androgen drugs, which have been extensively explored either alone or in combination with castration [2]. Flutamide (Eulexin®) (1) (in its active form as hydroxyflutamide (2)), bicalutamide (Casodex®) (3), nilutamide (Niladron®) (4) and enzalutamide (previously called MDV3100) (Xtandi®) (5) are all non-steroidal androgen receptor antagonists approved for the treatment of PC (Fig. 1). In many cases, after extended treatment over several years, these anti-androgens become ineffective and the disease may progress to a more aggressive and lethal form, known as castration resistant prostate cancer (CRPC). The major cause of this progressive disease is the emergence of different mutations on the AR, which cause the anti-androgen compounds to function as agonists, making them tumour-stimulating agents[3].

-
Fig. 1.
Structure of anti-androgen small molecules approved by FDA or in clinical development for the treatment of PC.
Among the drugs used for the treatment of PC, bicalutamide and enzalutamide selectively block the action of androgens while presenting fewer side effects in comparison with other AR antagonists [4], [5] and [6]. The structure of these molecules is characterised by the presence of a trifluoromethyl substituted anilide, which appears to be critical for biological activity (Fig. 1). As a means to improve the anti-proliferative activity of these compounds, and in order to exploit the well established potential of the fluorine atom in enhancing the pharmacological properties and drug-like physicochemical characteristics of candidate compounds [7], [8] and [9], a wide array of diverse new structures has been rationally designed and synthesised, through the introduction of fluoro-, trifluoromethyl- and trifluoromethoxy groups in diverse positions of both aromatic rings of the parent scaffolds. Our modifications resulted in a marked improvement of in vitro anti-proliferative activities on a range of human PC cell lines (VCap, LNCaP, DU-145 and 22RV1). In addition, we probed full versus partial AR antagonism for our new compounds.
Paper

Volume 118, 8 August 2016, Pages 230–243

Design and synthesis of novel bicalutamide and enzalutamide derivatives as antiproliferative agents for the treatment of prostate cancer
School of Pharmacy and Pharmaceutical Sciences, Redwood Building, King Edward VII Avenue, CF10 3NB, Cardiff, Wales, UK
- doi:10.1016/j.ejmech.2016.04.052
- E-mail address: Ferlas1@cardiff.ac.uk (Dr. S. Ferla)
Highlights
- •Synthesis of novel fluorinated bicalutamide and enzalutamide analogs.
- •Anti-proliferative activity in four human prostate cancer cell lines improved up to 50 folds.
- •Full AR antagonist effect exhibited by the new compounds.
- •Activity switch from partial agonist to full AR antagonist for enobosarm scaffold.
- •AR open conformation homology model and molecular modeling studies.
Abstract
Prostate cancer (PC) is one of the major causes of male death worldwide and the development of new and more potent anti-PC compounds is a constant requirement. Among the current treatments, (R)-bicalutamide and enzalutamide are non-steroidal androgen receptor antagonist drugs approved also in the case of castration-resistant forms. Both these drugs present a moderate antiproliferative activity and their use is limited due to the development of resistant mutants of their biological target.
Insertion of fluorinated and perfluorinated groups in biologically active compounds is a current trend in medicinal chemistry, applied to improve their efficacy and stability profiles. As a means to obtain such effects, different modifications with perfluoro groups were rationally designed on the bicalutamide and enzalutamide structures, leading to the synthesis of a series of new antiproliferative compounds. Several new analogues displayed improved in vitro activity towards four different prostate cancer cell lines, while maintaining full AR antagonism and therefore representing promising leads for further development.
Furthermore, a series of molecular modelling studies were performed on the AR antagonist conformation, providing useful insights on potential protein-ligand interactions.
http://www.sciencedirect.com/science/article/pii/S0223523416303452
Top cancer scientist dies of the disease he spent his life trying to cure
Professor Chris McGuigan, 57, of Cardiff University, was trying to invent new drugs to use in the fight against the disease

Professor Chris McGuigan, 57, was trying to invent new drugs to use in the fight against the disease.
But the tragic scientist, who was head of medicinal chemistry at Cardiff University’s School of Pharmacy and Pharmaceutical Sciences, died after his own fight with cancer.
A spokesman for Cardiff University said: “Professor McGuigan had been at the heart of scientific research for more than 30 years. He was an exceptionally gifted inventor and chemist.
“His loss will be felt cross the university and the wider scientific community.

“He had a strong drive to use his scientific ideas for social good, working tirelessly to address medical needs where they were unmet.
“Our thoughts are with his family, friends and close colleagues at this very sad time.”
Prof McGuigan’s research led him to try and develop new drugs for cancer, HIV, hepatitis B and C, shingles, measles, influenza and central nervous system (CNS) disease.
He also invented four new experimental drugs that were used in human clinical trials.
Prof McGuigan, who lived in Cardiff, is survived by wife Maria, 50, and his two young daughters Phoebe and Grace.
References
-
- Estimates of worldwide burden of cancer in 2008: GLOBOCAN 2008
- Int. J. Cancer, 127 (2010), pp. 2893–2917
-
- Antiandrogen monotherapy: a new form of treatment for patients with prostate cancer
- Urology, 58 (2001), pp. 16–22
-
- Flutamide withdrawal syndrome: its impact on clinical trials in hormone-refractory prostate cancer
- J. Clin. Oncol., 11 (1993), pp. 1566–1572
-
- Antiandrogens in prostate cancer
- Investig. New Drugs, 17 (1999), pp. 271–284
-
- The role of antiandrogen monotherapy in the treatment of prostate cancer
- BJU Int., 91 (2003), pp. 455–461
-
- Antiandrogens in the treatment of prostate cancer
- Eur. Urol., 51 (2007), pp. 306–313
-
- Fluorine-containing natural products
- J. Fluor. Chem., 100 (1999), pp. 127–133
-
- Fluorine substituent effects on bioactivity
- J. Fluor. Chem., 109 (2001), pp. 3–11
-
- Fluorine in pharmaceutical industry: fluorine-containing drugs introduced to the market in the last decade 2001–2011
- Chem. Rev., 114 (2014), pp. 2432–2506
-
- A two-step synthesis of the anti-cancer drug (R,S)-Bicalutamide
- Synthesis, 7 (2002), pp. 850–852
-
- Nucleohilic aromatic substitution of methacrylamide anion and its application to the synthesis of the anticancer drug bicalutamide
- J. Org. Chem., 26 (2003), pp. 10181–10182
-
-
- Bicalutamide: clinical pharmacokinetics and metabolism
- Clin. Pharmacokinet., 13 (2004), pp. 855–878
-
-
- Resolution of the nonsteroidal antiandrogen 4′-cyano-3-[(4-fluorophenyl)sulfonyl]-2-hydroxy-2-methyl-3′-(trifluoromethyl)-propionanilide and the determination of the absolute configuration of the active enantiomer
- J. Med. Chem., 31 (1988), pp. 885–887
-
- Novel nonsteroidal ligands with binding affinity and potent functional activity for the androgen receptor
- Eur. J. Med. Chem., 37 (2002), pp. 619–634
///////////1929605-82-2, bicalutamide and enzalutamide derivatives, antiproliferative agents, treatment of prostate cancer, School of Pharmacy and Pharmaceutical Sciences, Redwood Building, King Edward VII Avenue, CF10 3NB, Cardiff, Wales, UK
FC(F)(F)c1cc(ccc1[N+]([O-])=O)NC(=O)C(O)(CSc2ccccc2C(F)(F)F)C(F)(F)F
Eldecalcitol, an active vitamin D3 analog used to treat osteoporosis
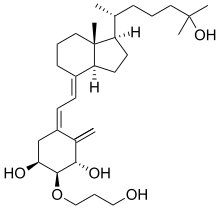
Eldecalcitol
(1S,2S,3S,5Z)-5-[(2E)-2-[(1R,3aS,7aR)-1-[(2R)-6-hydroxy-6-methylheptan-2-yl]-7a-methyl-2,3,3a,5,6,7-hexahydro-1H-inden-4-ylidene]ethylidene]-2-(3-hydroxypropoxy)-4-methylidenecyclohexane-1,3-diol
(1R,2R,3R,5Z,7E)-2-(3-Hydroxypropyloxy)-9,10-secocholesta-5,7,10(19)-triene-1,3,25-triol
| AC1O5QQ2; CAS 104121-92-8; AN-3697; ED 71, Edirol® | |
| Molecular Formula: | C30H50O5 |
|---|---|
| Molecular Weight: | 490.715 g/mol |
APPROVED JAPAN , 2011-01-21, Chugai (Originator) , Roche,Taisho Toyama
Eldecalcitol was approved by Pharmaceuticals and Medicals Devices Agency of Japan (PMDA) on January 21, 2011. It was developed by Chugai Pharmaceutical (a member of Roche) and marketed as Edirol® by Chugai Pharmaceutical and Taisho.
Eldecalcitol is an orally active vitamin D analogue leading to greater absorption of bind calcium. It is usually used to treat osteoporosis.
Edirol® is available as capsule for oral use, containing 0.5 μg or 0.75 μg of free Eldecalcitol, and the recommended dose is 0.75 μg once daily.
ED-71, a vitamin D analog, is a more potent inhibitor of bone resorption than alfacalcidol in an estrogen-deficient rat model of osteoporosis. ED-71, effectively and safely increased lumbar and hip bone mineral density (BMD) in osteoporotic patients who also received vitamin D3 supplementation.
Eldecalcitol is a drug used in Japan for the treatment of osteoporosis.[1] It is an analog of vitamin D.[2] Osteoporosis is a common bone disease among the older generation, with an estimated prevalence of over 200 million people.[1] This condition often results in bone fractures due to abnormally low bone mass density, and is a leading cause of disability, especially among developed countries with longer average life spans. Osteoporosis is more common in women than with men.
Discovery
Chugai Pharmaceutical/Roche are the originators of the medicinal drug eldecalcitol through Taisho Pharmaceutical Holdings and Chugai Pharmaceutical. The trade name of eldecalcitol is Edirol, and its Chemical Abstracts Service (CAS) registry number is 104121-92-8. Eldecalcitol was approved for use in Japan on January 2011. The approval came from the Japanese Ministry of Health, Labor, and Welfare for the objective of a treatment for osteoporosis.[3]
Effects
Clinical trials have suggested that eldecalcitol, a vitamin D analog, has strong effects to reduce calcium reabsorption into the body from bones, therefore increasing bone mineral density, and to increase calcium absorption in intestines.[4] In animals, eldecalcitol inhibits the activity of osteoclasts for the function to reduce bone degradation for calcium, while still able to maintain osteoblast function so as to not hinder bone formation.[5] Unlike other vitamin D analogs, eldecalcitol does not significantly suppress parathyroid hormone levels, promising a better treatment for osteoporosis in comparison to other medications.[6] Bone mineral density increases with eldecalcitol use, in addition to strengthening bone structure. This occurs due to the function of the eldecalcitol drug, which decreases bone reabsorption as observed through a bone reabsorption marker. Bone geometry assessments show that eldecalcitol increases cortical bone area in patients with osteoporosis more so than other vitamin D analogs, such as alfacalcidol. There was also the maintenance of thickness of cortical bone mass, strongly indicating that eldecalcitol improves the strength and mass of bone, specifically cortical bone structure.[7] Adverse effects of eldecalcitol include an increase in blood and urinary calcium levels. Abnormally high levels of calcium can lead to problems associated with hypercalcemia.
Treatment for Osteoporosis
Eldecalcitol can be used for the treatment of hypocalcaemia or osteoporosis. Calcium absorption increases with the presence of eldecalcitol by the body, occurring in the intestines, which is useful for those who have low calcium levels. Eldecalcitol is more often used due to its effects to treat osteoporosis. In the aging population, the bone matrix becomes weakened through untreated osteoporosis. This leads to an increased risk of severe fractures that include spinal and hip fractures in addition to vertebral and wrist fractures. This creates a burden on the health care system due to a decline in the quality of life for the individuals that suffer from this condition. Some risk factors leading to the predisposition of developing osteoporosis are previous incidents of bone fractures and a reduction in bone mineral density.[1] These factors expectantly increase as age increases. Bone health is reliant on maintaining physiologically needed levels of calcium, where the body constantly maintains this calcium homeostasis through osteoblast and osteoclast activity. Osteoblast activity serves this function of maintaining appropriate calcium levels by depositing calcium in bones when blood calcium levels are above normal. In contrast, osteoclasts break down bone tissue to increase blood calcium levels if they are low.[8] This activity is performed after absorption of calcium by the body, which requires the actions of vitamin D. The active metabolite of vitamin D, calcitriol, performs its function through interactions with the calcitriol receptor. This nuclear hormone receptor is responsible for calcium absorption which, in turn, is involving in bone depletion and formation. The new analogs of vitamin D, such as eldecalcitol, are observed to have stronger effects in preventing bone loss, fractures, and falls in comparison to calcitriol.[9] Eldecalcitol is even more effective than its counterpart alfacalcidol, another vitamin D analog. Studies have shown eldecalcitol is more effective than alfacalcidol in preventing vertebral and wrist fractures, and even falls, with osteoporotic patients with vitamin D insufficiencies.[10] Eldecalcitol is also more effective at preventing fractures than vitamin D and calcium supplements.[1] Eldecalcitol increases calcium absorption for vitamin D deficient patients, and therefore could be used for osteoporosis treatment for all age groups.
Pharmacology
Analogs of vitamin D are being explored intensely for their regulatory effects on calcium metabolism with the purpose of treating osteoporosis, a skeletal disease associated with low bone mass and deterioration of bone tissue. Vitamin D is imperative for absorption of calcium to maintain bone strength.
Mechanism of Action
Eldecalcitol is an orally administered drug to patients, which binds to vitamin D receptors and binding protein for the goal of achieving greater specificity to bind calcium for its absorption. This greater affinity is 2.7-fold that of the active vitamin D form of calcitriol. Eldecalcitol is readily absorbed into the body, with a long elimination half-life of over eight hours, reaching maximum absorption in 3.4 hours.[1]
Dosage
Eldecalcitol is present in the form of pills for oral administration. In preclinical models with healthy male volunteers, oral doses of eldecalcitol ranged from 0.1 to 1.0 micrograms once daily to show an increase in bone mineral density.[11] Preclinical trials show improvements for doses at 0.5 and 0.75 micrograms, which are the recommended dosage amounts for the Edirol product as approved by the Japanese Ministry of Health, Labor, and Welfare for treating osteoporosis.[3]
Chemistry
The class of eldecalcitol is a vitamin D3 derivative. This molecule has a molecular weight of 490.71 grams per mole. The eldecalcitol analog of calcitriol, contains a hydroxypropyl group in the lower cyclohexane ring. The synthesis of eldecalcitol incorporates two units assembled together. The IUPAC names include (3S, 4S, 5R)-oct-1-en-7-yne-3,4,5-triol that is fused to a bicyclic system, (R)-6-((1R, 3aR, 7aR, E)-4-(bromomethylene)-7a-methyloctahydro-1H-inden-1-yl)-2-methylheptan-2-ol. The assembly process includes a Diels-Alder reaction to give the fully protected eldecalcitol. In order to get the parent molecule, the hydroxyl groups have to be deprotected. The chemistry of eldecalcitol allows for its binding 2.7-fold more potently than calcitriol. In addition, some vitamin D derivatives have been known to inhibit the serum parathyroid hormone. Eldecalcitol only weakly inhibits the serum parathyroid hormone, making it an even more appealing medicinal drug for its physiological uses in the treatment of osteoporosis.[3] Animal studies of eldecalcitol, in ovariectomized rats, show improvements in bone mass while lowering bone reabsorption to demonstrate its effectiveness in osteoporosis treatment.[5]
PAPER
Heterocycles, Vol 92, No. 6, 2016, pp.1013-1029
Published online, 22nd March, 2016
■ Diverse and Important Contributions by Medicinal Chemists to the Development of Pharmaceuticals: An Example of Active Vitamin D3 Analog, Eldecalcitol
Noboru Kubodera*
*International Institute of Active Vitamin D Analogs, 35-6, Sankeidai, Mishima, Shizuoka 411-0017, Japan
Abstract
Presented herein are diverse and important contributions by medicinal chemists to different stages of pharmaceutical development. The conceptual elements reviewed, which are intended for young chemists who engage in drug discovery research, draw upon the author’s experience in developing eldecalcitol, an active vitamin D3 analog used to treat osteoporosis. The review covers exploratory research for a lead candidate compound; process development for practical manufacturing; and synthesis of other compounds relevant to the program, such as tritiated compounds, postulated metabolites, and miscellaneous analogs for mode of action studies.
PAPER
Eldecalcitol [1α,25-dihydroxy-2β-(3-hydroxypropoxy)vitamin D3], an analog of calcitriol (1α,25-dihydroxyvitamin D3), possesses a hydroxypropoxy substituent at the 2β-position of calcitriol. Eldecalcitol has potent biological effects on bone disease such as osteoporosis. The marketing of eldecalcitol has very recently started in Japan. In consideration of this, we have been investigating practical synthesis of eldecalcitol for industrial-scale production. Eldecalcitol was initially synthesized in a linear manner. The 27-step linear sequence was, however, suboptimal due to its lengthiness and low overall yield (ca. 0.03%). Next, we developed a convergent approach based on the Trost coupling reaction, in which the A-ring fragment (ene-yne part obtained in 10.4% overall yield) and the C/D-ring fragment (bromomethylene part obtained in 27.1% overall yield) are coupled to produce the triene system of eldecalcitol (15.6%). Although the overall yield of the convergent synthesis was better than that of the linear synthesis, significant improvements were still necessary. Therefore, additional biomimetic studies were investigated. Process development for the practical production of eldecalcitol is described herein.
http://ar.iiarjournals.org/content/32/1/303/F3.expansion.html
Convergent synthesis of eldecalcitol (5) by coupling A-ring fragment 37 with C/D-ring fragment 40. Reagents and conditions: a: HO(CH2)3OH/t-BuOK, 120°C. b: t-BuCOCl/pyridine/CH2Cl2, rt. c: H2/Pd(OH)2/MeOH, rt. d: Me2C(OMe)2/TsOH/acetone, rt. e: DMSO/(COCl)2/CH2Cl2, −60°C. f: CH2=CHMgBr/THF, −60°C. g: t-BuCOCl/Et3N/DMAP/CH2Cl2, rt. h: 1 M HCl/MeOH, rt. i: Ph3P/DEAD/benzene, reflux. j: LiC ≡ CTMS/BF3-OEt2, −78°C. k: 10 N NaOH/MeOH, rt. l: TBSOTf/Et3N/CH2Cl2, 0°C. m: TESOTf/Et3N/CH2Cl2, 0°C. n: O3/CH2Cl2/MeOH, −78°C then NaBH4/MeOH, −78°C. o: NMO/TPAP/4Ams/CH2Cl2, rt. p: Ph3P+CH2BrBr−/NaHMDS/ THF, −60°C to rt. q: (dba)3Pd2-CHCl3/PPh3/Et3N/toluene, reflux. r: TBAF/THF/toluene, reflux.

Industrial synthesis of alfacalcidol (4) and biomimetic synthesis of eldecalcitol (5) from cholesterol (42). Reagents and conditions: a: [Al(Oi-Pr)3]/cyclohexanone. b: DDQ/AcOEt. c: NaOEt/EtOH. d: NaBH4/MeOH/THF. e: Ac2O/DMPA/pyridine, rt. f: NBS/AIBN/n-hexane, reflux. g: γ-collidine/toluene, reflux. h: KOH/MeOH, rt. i: PTAD/CH2Cl2, rt. j: TBSCl/imidazole. k: MCPBA/CH2Cl2. l: DMI, 140°C. m: TBAF/THF. n: NaBH4/EtOH. o: 400 W high pressure mercury lamp/THF, 0°C then reflux without mercury lamp. p: HO(CH2)3OH/t-BuOK, 110°C. q: Microbial 25-hydroxylation.

ROUTE1
Reference:1. US4666634A / EP184206A2.

Reference:1. Anticancer. Res. 2012, 32, 303-310.
2. Drugs. Fut. 2005, 30, 450-461.

Reference:1. J. Am. Chem. Soc. 2001, 123, 3716-3722.

Reference:1. Bioorg. Med. Chem. Lett. 1997, 7, 2871-2874.
2. Anticance. Res. 2009, 29, 3571-3578.

Reference:1. EP0503630A1.
2. Drugs Fut. 2005, 30, 450-461.

Reference:1. Bioorg. Med. Chem. 1998, 6, 2517-2523.
References
- Sanford, M; McCormack, PL (2011). “Eldecalcitol: A review of its use in the treatment of osteoporosis”. Drugs 71 (13): 1755–70. doi:10.2165/11206790-000000000-00000. PMID 21902297.
- Hatakeyama, S; Yoshino, M (2010). “Synthesis and preliminary biological evaluation of 20-epieldecalcitol [20-epi-1α,25-dihydroxy-2β-(3-hydroxypropoxy)vitamin D3: 20-epi-ED-71]”. The Journal of Steroid Biochemistry and Molecular Biology 121 (1–2): 25–28.doi:10.1016/j.jsbmb.2010.03.041. PMID 20304058.
- Robichaud; Stamford; Weinstein; McAlpine; Primeau; Lowe; Bernstein; Bronson; Manoj, Desai (2012). Annual Reports in Medicinal Chemistry 47 (1st ed.). San Diego: Elsevier Inc. pp. 529–531. ISBN 9780123964922.
- Nogachi, Y; Kawate, H; Nomura, M; Takayanagi, R (2013). “Eldecalcitol for the treatment of osteoporosis”. Europe PubMed Central 8: 1313–1321. doi:10.2147/CIA.S49825.
- Smith, S; Doyle, N; Boyer, M; Chouinard, L; Saito, H (2013). “Eldecalcitol, a vitamin D analog, reduces bone turnover and increases trabecular an cortical bone mass, density, and strength in ovariectomized cynomolgus monkeys”. Bone 57 (1): 116–122.doi:10.1016/j.bone.2013.06.005. PMID 23774444.
- Harada, S; Uno, S; Takahashi, F; Saito, H (2010). “Eldecalcitol is less effective in suppressing parathyroid hormone compared to calcitriol in vivo“. The Journal of Steroid Biochemistry and Molecular Biology 121 (1–2): 281–283.doi:10.1016/j.jsbmb.2010.04.001. PMID 20398764.
- Nakamura, T; Takano, T; Fukunaga, M; Shiraki, M; Matsumoto, T (2013). “Eldecalcitol is more effective for the prevention of osteoporotic fractures than alfacalcidol”. Journal of Bone and Mineral Metabolism 31 (4): 417–422. doi:10.1007/s00774-012-0418-5.PMC 3709079. PMID 23575909.
- Matsuo, K; Irie, N (2008). “Osteoclast-osteoblast communication”. Archives of Biochemistry and Biophysics 473 (2): 201–209. doi:10.1016/j.abb.2008.03.027.PMID 18406338.
- Saito, H; Takeda, S; Amizuka, N (2013). “Eldecalcitol and calcitriol stimulates ‘bone minimodeling,’ focal bone formation without prior bone resorption, in rat trabecular bone”.The Journal of Steroid Biochemistry and Molecular Biology 136 (1): 178–182.doi:10.1016/j.jsbmb.2012.10.004.
- Matsumoto, T; Ito, M; Hayashi, Y; Hirota, T; Tanigawara, Y; Sone, T; Fukunaga, M; Shiraki, M; Nakamura, T (2011). “A new active vitamin D3 analog, eldecalcitol, prevents the risk of osteoporotic fractures—A randomized, active comparator, double-blind study”. Bone49 (4): 605–612. doi:10.1016/j.bone.2011.07.011. PMID 21784190.
- Harada, S; Mizoguchi, T; Kobayashi, Y; Nakamichi, Y; Takeda, S; Sakai, S; Takahashi, F; Saito, H; Yasuda, H; Udagawa, N; Suda, T; Takahashi, N (2012). “Daily administration of eldecalcitol (ED-71), an active vitamin D analog, increases bone mineral density by suppressing RANKL expression in mouse trabecular bone”. Journal of Bone and Mineral Research 27 (1): 461–473. doi:10.1002/jbmr.555.
| No. | Major Technical Classification | Publication No. | Patent No. | Legal Status | Filling Date | Estimated Expiry Date |
| 1 | Preparation | CN85108857A | CN1008368B | Granted/expired | 1985/12/4 | 2005/12/4 |
| 2 | Crystal | CN1223639A | CN1216861C | Granted | 1997/6/16 | 2017/6/16 |
| 3 | Preparation | CN1637017A | CN1276927C |
| Patent ID | Date | Patent Title |
|---|---|---|
| US7927613 | 2011-04-19 | Pharmaceutical co-crystal compositions |
| US7323580 | 2008-01-29 | CRYSTALS OF A VITAMIN D DERIVATIVE AND A METHOD FOR THE PREPARATION THEREOF |
| US7235679 | 2007-06-26 | Crystals of a vitamin D derivative and a method for the preparation thereof |
| EP0924199 | 2006-05-10 | CRYSTALS OF VITAMIN D DERIVATIVES AND PROCESS FOR THE PREPARATION THEREOF |
| US2005009794 | 2005-01-13 | Crystals of a vitamin D derivative and a method for the preparation thereof |
| US6831183 | 2004-12-14 | Crystals of a vitamin D derivative and a method for the preparation thereof |
| US6448421 | 2002-09-10 | CRYSTALS OF VITAMIN D DERIVATIVES AND PROCESS FOR THE PREPARATION THEREOF |
 |
|
| Systematic (IUPAC) name | |
|---|---|
|
(1S,2S,3S,5Z,7E)-2-(3-Hydroxypropoxy)-9,10-secocholesta-5,7,10-triene-1,3,25-triol
|
|
| Clinical data | |
| Trade names | Edirol |
| Identifiers | |
| CAS Number | 104121-92-8 |
| ATC code | None |
| PubChem | CID 6438982 |
| ChemSpider | 4943418 |
| Chemical data | |
| Formula | C30H50O5 |
| Molar mass | 490.715 g/mol |
///////////eldecalcitol, active vitamin D3 analog, treat osteoporosis, AC1O5QQ2, 104121-92-8, AN-3697, ED 71, ED-71, Edirol®, PMDA, JAPAN
O[C@H]1CC(\C(=C)[C@H](O)[C@H]1OCCCO)=C\C=C2/CCC[C@]3([C@H]2CC[C@@H]3[C@H](C)CCCC(O)(C)C)C
OR
CC(CCCC(C)(C)O)C1CCC2C1(CCCC2=CC=C3CC(C(C(C3=C)O)OCCCO)O)C
MK 8876
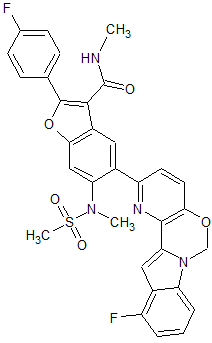

MK 8876
CAS 1426960-33-9
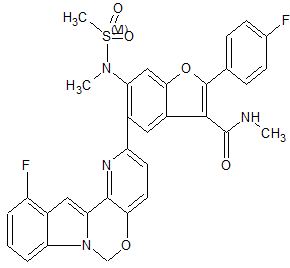
2-(4-Fluorophenyl)-5-(11-fluoro-6H-pyrido[2′,3′:5,6][1,3]oxazino[3,4-a]indol-2-yl)-N-methyl-6-(N-methylmethanesulfonamido)-1-benzofuran-3-carboxamide
| 2-(4-Fluorophenyl)-5-(11-fluoro-6H-pyrido[2′,3′:5,6][1,3]oxazino[3,4-a]indol-2-yl)-N-methyl-6-[methyl(methylsulfonyl)amino]-3-benzofurancarboxamide |
| Molecular Formula | C32H24F2N4O5S | |
| Molecular Weight | 614.62 |
- Originator Merck & Co
- Class Antivirals
- Phase I Hepatitis C
Most Recent Events
- 11 Oct 2013 Phase-I clinical trials in Hepatitis C in Germany (PO)
- 11 Oct 2013 Phase-I clinical trials in Hepatitis C in Moldova (PO)
- 23 Aug 2013 Preclinical trials in Hepatitis C in USA (PO)
DATA
2-(4-Fluorophenyl)-5-(11-fluoro-6H-pyrido[2′,3′:5,6][1,3]oxazino[3,4-a]indol-2-yl)-N-methyl-6-(N-methylmethanesulfonamido)-1-benzofuran-3-carboxamide
MK-8876 off-white solid
1H NMR (500 MHz, DMSO-d6) δ 8.56 (q, J = 4.7 Hz, 1H), 8.06–8.01 (m, 2H), 8.05 (s, 1H), 7.86 (s, 1H), 7.71 (d, J = 8.5 Hz, 1H), 7.62 (d, J = 8.5 Hz, 1H), 7.52 (d, J = 8.3 Hz, 1H), 7.46–7.40 (m, 2H), 7.29–7.22 (m, 1H), 7.11 (s, 1H), 6.94 (dd, J = 10.6, 7.9 Hz, 1H), 6.27 (s, 2H), 3.31 (s, 3H), 2.96 (s, 3H), 2.85 (d, J = 4.7 Hz, 3H);
13C NMR (125.7 MHz, DMSO-d6) δ 162.86, 162.82 (d, JC–F = 248.5 Hz), 155.74 (d, JC–F = 246.1 Hz), 153.80, 152.43, 152.28, 147.20, 137.08, 137.00 (d, JC–F = 10.8 Hz), 136.36, 136.20, 132.37, 129.50 (d, JC–F = 8.6 Hz), 127.17, 125.45 (d, JC–F = 3.1 Hz), 125.08, 125.02, 123.70 (d, JC–F = 7.7 Hz), 122.28, 117.23 (d, JC–F = 22.4 Hz), 116.01 (d, JC–F = 21.9 Hz), 113.65, 111.76, 106.90 (d,JC–F = 3.5 Hz), 105.32 (d, JC–F = 18.5 Hz), 94.16, 73.57, 39.39, 37.24, 26.16;
HR-ESI-MS m/zcalcd for C32H25N4O5SF2+ [M + H]+ 615.1514, found 615.1500.
| HPLC Method | |
|---|---|
| column | Ascentis Express C18 2.7 μm (fused core), 100 mm × 4.6 mm |
| detection | UV at 210 nm |
| column temperature | 40 °C |
| flow rate | 1.8 mL/min |
| injection volume | 5.0 μL |
| gradient | 90% A to 5% A over 11 min, hold at 5% A for 2 min, 5% A back to 90% A over the next 0.1 min, and then hold at 90% A for 2.9 min |
| run time | 16 min |
| data collection | acquisition for the first 13 min |
| mobile phases | solvent A: water with 0.1% H3PO4 |
| solvent B: acetonitrile | |
| Retention Time Data | |
|---|---|
| identity | tR (min) |
| boronic acid 27 | 4.24 |
| desbromoarene 28 | 5.33 |
| MK-8876 (1) | 7.89 |
| chloropyridine starting material 2 | 8.03 |
| BHT | 10.22 |
SYNTHESIS


CONTD……………
MK 8876


MK 8876
Patent
Scheme 1

Scheme 2

Scheme 3

Q
Scheme 4

EXAMPLES
Example 1
Preparation of Compound 1
 THIS COMPD HAS ONE FLUORO MISSING, APPLY TO YOUR MK 8876
Step 1 – Synthesis of 2,6-dichloropyridin-3-ol

Η202 (1.60 g, 47.12 mmol) was added slowly to the solution of compound 2,6- dichloropyridin-3-ylboronic acid (3 g, 15.71 mmol) in CH2CI2 (30 mL) at 0 °C. After stirred at room temperature for about 15 hours, the mixture was quenched with sat. Na2S203 aqueous (50 mL) and adjusted to pH < 7 with IN HC1. The mixture was extracted with EtOAc (40 mL x 3). The organic layer was washed with brine (100 mL), dried over Na2S04, filtered and the solvent was evaporated to provide2,6-dichloropyridin-3-ol (2.34 g, yield: 91.4%). 1H-NMR (CDC13, 400 MHz) δ 7.30 (d, / = 8.4 Hz, 1H), 7.19 (d, / = 8.4 Hz, 1H), 5.70 (br, 1H).
– Synthesis of 2,6-dichloro- -methoxypyridine

To a solution of 2,6-dichloropyridin-3-ol (16.3 g, 0.1 mol) and K2C03 (41.4 g, 0.3 mol) in DMF (200 mL) were added Mel (21.3 g, 0.15 mol). The mixture was allowed to stir at 80 °C for 2 hours. The mixture was then diluted with water (200 mL) and extracted with EtOAc (200 mL x 3). The organic layer was washed with brine (200 mL x 3), dried over Na2S04, filtered and the solvent was evaporated to provide 2,6-dichloro-3-methoxypyridine (17.0 g, yield: 96.0%). 1H-NMR (CDC13, 400 MHz) δ 7.12-7.18 (m, 2H), 3.86 (s, 3H). Step 3 – Synthesis of2-(6-chloro-3-methoxypyridin-2-yl)-lH-indole

To a degassed solution of compound 2,6-dichloro-3-methoxypyridine (8.9 g, 0.05 mol), (l-(tert-butoxycarbonyl)-lH-indol-2-yl)boronic acid (13 g, 0.05 mol) and K3PO4 (31.8 g, 3.0 mol) in DMF (100 mL) was added Pd(dppf)Cl2 (3.65 g, 0.005 mol) under N2. The mixture was heated at 60 °C for about 15 hours. The reaction mixture was cooled to room temperature, diluted with EtOAc and filtered. The filtrate was washed with H20, brine, dried over Na2S04. After being concentrated in vacuo, the resulting residue was purified using prep-HPLC to provide the desired product of 2-(6-chloro-3-methoxypyridin-2-yl)-lH-indole (9.0 g, yield:
69.8%). 1H-NMR (CDC13, 400 MHz) δ 9.52 (s, 1H), 7.65 (d, / = 7.6 Hz, 1H), 7.38-7.43 (m, 2H), 7.07-7.26 (m, 4H), 4.03 (s, 3H).
Step 4 – Synthesis of6-chlor -2-(lH-indol-2-yl)pyridin-3-ol

BBr3 (0.4 mL, 0.39 mmol) was added to the solution of 2-(6-chloro-3- methoxypyridin-2-yl)-lH-indole (50 mg, 0.194 mmol) in CH2C12 (0.5 mL) at -78 °C under N2. The mixture was allowed to stir at room temperature for 3 hours. The mixture was then quenched with CH3OH (10 mL) at -78 °C. After being concentrated in vacuo, the resulting residue was purified using prep-TLC (PE : EtOAc = 2.5 : 1) to afford the desired product of 6- chloro-2-(lH-indol-2-yl)pyridin-3-ol (40 mg, yield: 85.1%). 1H-NMR (CDC13, 400 MHz) δ 10.09 (s, 1H), 9.72 (s, 1H), 7.50 (d, / = 7.9 Hz, 1H), 7.17-7.32 (m, 3H), 7.08-7.14 (m, 1H), 6.87-6.96 (m, 2H).
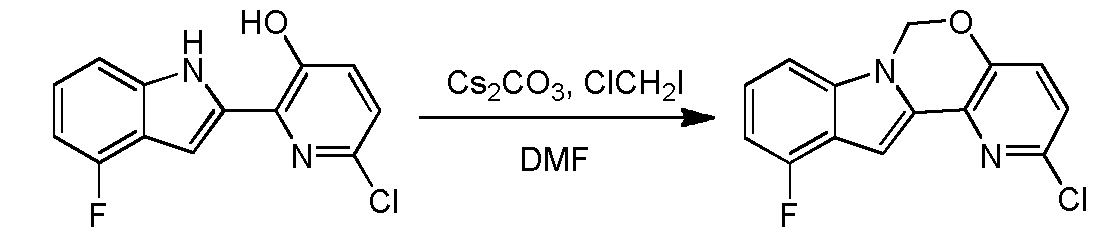
Step 5 – Synthesis of 2-chlo -6H-pyrido[2′ ,3′ : 5 ,6] [ 1 ,3]oxazino[3 ,4-a]indole

To a solution of chloroiodomethane (3.51 g, 20.0 mmol) and K2CO3 (1.38 g, 10.0 mmol) in DMF (50 mL) was allowed to stir at 100 °C, 6-chloro-2-(lH-indol-2-yl)pyridin-3-ol (480 mg, 2.0 mmol) in DMF (50 mL) was added dropwise. After addition, the mixture was allowed to stir for another 0.5 hours. The mixture was then diluted with water (100 mL) and extracted with EtOAc (100 mL x 3). The organic layer was washed with brine (100 mL x 3), dried over Na2S04 and concentrated. The residue was purified using prep-TLC (PE : EtOAc = 3 1) to afford the desired product of 2-chloro-6H-pyrido[2′,3′:5,6][l,3]oxazino[3,4-a]indole (260 mg, yield: 50.7%). 1H-NMR (CDC13, 400 MHz) δ 7.63 (d, / = 8.0 Hz, 1H), 7.22-7.27 (m, 3H), 7.19 (d, / = 2.4 Hz, 1H), 7.08-7.12 (m, 2H), 5.86 (s, 2H).
Step 6 – Synthesis of2-(4-fluowphenyl)-N-methyl-6-(N-methylmethylsulfonamido)-5-(6H- pyridol 2 ‘,3’:5,6][ l, mpound 1 )
 THIS COMPD HAS ONE FLUORO MISSING, APPLY TO YOUR MK 8876
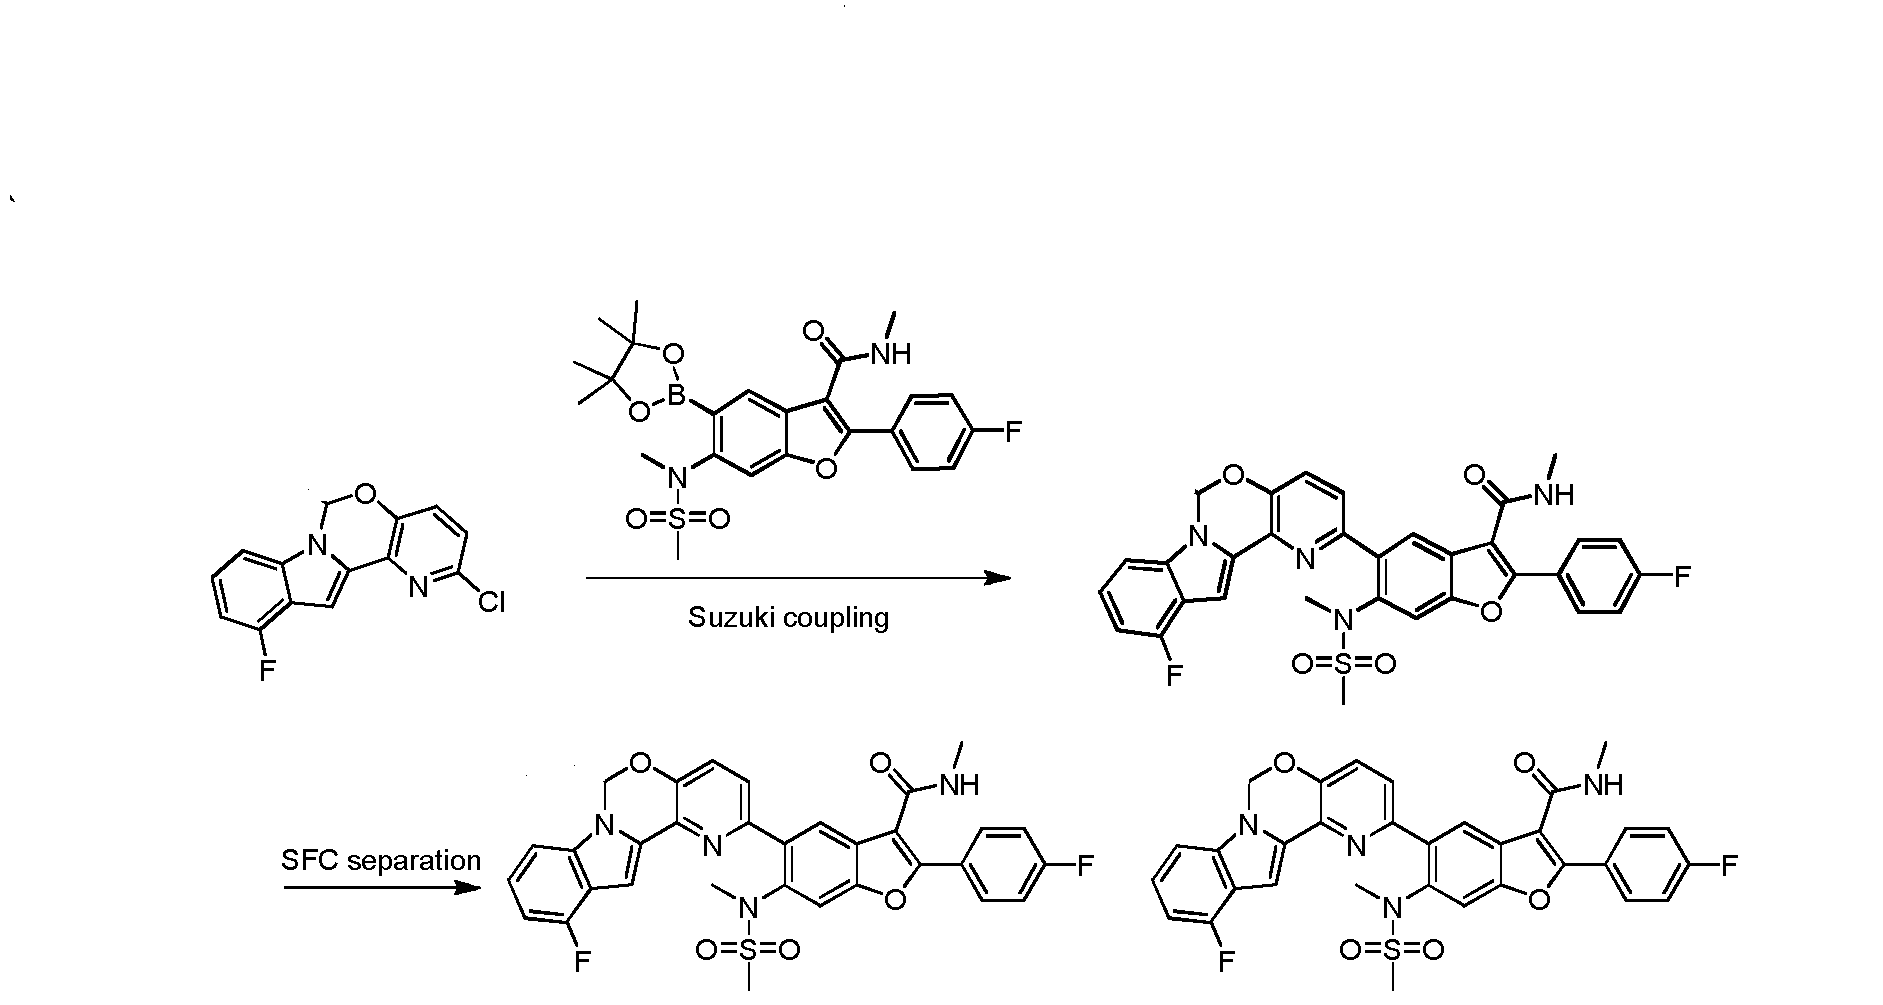
To a degassed solution of 2-(4-fluorophenyl)-N-methyl-6-(N- methylmethylsulfonamido)-5-(4,4,5,5-tetramethyl-l,3,2-dioxaborolan-2-yl)benzofuran-3- carboxamide (502 mg, 1.0 mmol), 2-chloro-6H-pyrido[2′,3′:5,6][l,3]oxazino[3,4-a]indole (256 mg, 1.0 mmol) and K3PO4 (636 mg, 3.0 mmol) in dioxane : H20 (1.5 mL : 0.4 mL) was added Pd2(dba)3 (91 mg, 0.1 mmol) and X-phos (91 mg, 0.2 mmol) under N2. The mixture was heated to 110 °C for 3 hours. The reaction mixture was cooled to room temperature, diluted with EtOAc and filtered. The filtrate was washed with H20, brine, dried over Na2S04. After being concentrated in vacuo, the resulting residue was purified using prep-HPLC to provide the desired product of Compound 1 (275 mg, yield: 46.1%). 1H-NMR (CDC13, 400 MHz) δ 7.88-7.94 (m, 3H), 7.61-7.63 (m, 2H), 7.40 (s, 2H), 7.09-7.28 (m, 6H), 5.94 (s, 2H), 5.86 (d, / = 4.4 Hz, 1H), 3.29 (s, 3H), 2.92 (d, / = 5.2 Hz, 3H), 2.65 (s, 3H). MS (M+H)+: 596.
Compounds 2-15, depicted in the table below, were prepared using the method described above.
COMPD 2 IS MK 8876

PATENT
Example 81
Preparation of Compound 2
Synthesis of ethyl 3- 4-fluorophenyl)-3-oxopropanoate
Diethyl carbonate (130 g, 1.1 mol) was dissolved in a suspension ofNaH (60% in oil, 50.2 g, 1.3 mol) in anhydrous tetrahydrofuran (1.5 L), and then l-(4-fluorophenyl)ethanone (150 g, 1.09 mol) was added dropwise at 70 °C. The resulting mixture was stirred at 70 °C for 3 hours. After the reaction mixture was cooled to room temperature and poured into HCl (1 N). The mixture was extracted with EtOAc, the organic phase was dried with anhydrous NaS04 and concentrated in vacuo. The resulting residue was purified using column chromatography (eluted with petroleum ether / EtOAc = 50 / 1) to provide ethyl 3-(4-fluorophenyl)-3-oxopropanoate (217 g, yield: 95%). 1H-NMR (CDC13, 400 MHz) δ 7.92-7.97 (m, 2H), 7.07-7.13 (m, 2H), 4.14-4.20 (m, 2H), 3.93 (s, 2H), 1.22 (d, J= 7.2 Hz, 3H). MS (M+H)+: 211. Step 2 – Synthesis of ethyl 5-bromo-2-(4-fluorophenyl)benzofuran-3-carboxylate
A solution of ethyl 3-(4-fluorophenyl)-3-oxopropanoate (130 g, 0.6 mol), 4- bromophenol (311 g, 1.8 mol) and FeCl3-6H20 (19.5 g, 0.09 mol) in DCE (700 mL) was heated to reflux, and then 2-(tert-butylperoxy)-2-methylpropane (193 g, 1.32 mol) was added dropwise under nitrogen. After 6 hours of refluxing, the mixture was cooled to RT, quenched with saturated NaHS03 and extracted with dichloromethane. The organic phases were washed with water, brine and dried over Na2S04, filtered and concentrated in vacuo. The resulting residue was purified using column chromatography (petroleum ether / dichloromethane = 15 / 1) to provide the crude product, which was crystallized from cold MeOH to provde ethyl 5-bromo-2- (4-fluorophenyl)benzofuran-3-carboxylate (37 g, yield: 14.3%) as solid. 1H- MR (CDC13, 400 MHz) δ 8.12 (s, 1H), 7.97-8.01 (m, 2H), 7.37 (d, J= 4.0 Hz, 1H), 7.32 (d, J= 8.0 Hz, 1H), 7.11 (t, J= 8.0 Hz, 2H), 4.32-4.38 (m, 2H), 1.36 (t, J= 8.0 Hz, 3H). MS (M+H)+: 363 / 365.
Step 3 – Synthesis of eth l 5-bromo-2-(4-fluorophen -6-nitrobenzofuran-3-carboxylate
To a solution of ethyl 5-bromo-2-(4-fluorophenyl)benzofuran-3-carboxylate (50 g,
137.6 mmol) in CHC13 (500 mL), fuming HN03 (50 mL) was added dropwise at -15 °C and the mixture was stirred for 0.5 hour. The reaction mixture was poured into ice water and extracted with CH2C12. The organic layer was washed with a.q. sat. NaHC03 and brine, after removed the most of solvent, the resulting residue was crystallized with petroleum ether / dichloromethane = 20 / 1 to provide product of ethyl 5-bromo-2-(4-fluorophenyl)-6-nitrobenzofuran-3-carboxylate (35 g, yield: 66%). 1H- MR (CDC13, 400 MHz) δ 8.36 (s, 1H), 8.02-8.04 (m, 3H), 7.13-7.18 (m, 2H), 4.36-4.41 (m, 2H), 1.37 (t, J= 4.0 Hz, 3H). MS (M+H)+: 408 / 410.
Step 4 – Synthesis of ethyl 6-amino-5-bromo-2-(4-fluorophenyl)benzofuran-3-carboxylate
A mixture of ethyl 5-bromo-2-(4-fluorophenyl)-6-nitrobenzofuran-3-carboxylate (52 g, 127 mmol), iron filings (21.3 g, 382.2 mmol) and H4C1 (41 g, 764.4 mmol) in MeOH / THF / H20 (2 / 2 / 1, 500 mL) was stirred at reflux for 3 hour. After filtered and concentrated, the resulting residue was purified using column chromatography (petroleum ether / EtOAc / dichloromethane = 20 : 1 : 20) to provide ethyl 6-amino-5-bromo-2-(4-fluorophenyl) benzofuran-3-carboxylate (40 g, yield: 82%). 1H- MR (CDC13, 400 MHz) δ 8.01 (s, 1H), 7.94-7.98 (m, 2H), 7.08 (t, J= 8.0 Hz, 2H), 6.83 (s, 1H), 4.32-4.36 (m, 2H), 4.18 (s, 2H), 1.35 (t, J= 8.0 Hz, 3H). MS (M+H)+: 378 / 380.
Step 5 – Synthesis of 5-Bromo-2-(4-fluoro-phenyl)-6-methanesulfonylamino-benzofuran-3- carboxylic acid eth l ester
MsCI (31.7 g, 277.5 mmol) was added to a solution of ethyl 6-amino-5-bromo-2- (4-fluorophenyl)benzofuran-3-carboxylate (35 g, 92.5 mmol) and pyridine (60 mL) in
dichloromethane (300 mL) at 0 °C. After stirred overnight at room temperature, the mixture was diluted with water and extracted with dichloromethane. The organic layer was washed with brine, dried over Na2S04, filtered and concentrated in vacuo, the resulting residue was purified using crystallized with EtOAc to provde the pure product of ethyl 5-bromo-2-(4-fluorophenyl)-6- (methylsulfonamido)benzofuran-3-carboxylate (35 g, yield: 82%). 1H- MR (CDC13, 400 MHz) δ 8.27 (s, 1H), 8.01-8.05 (m, 2H), 7.87 (s, 1H), 7.15-7.19 (m, 2H), 6.87 (s, 1H), 4.38-4.43 (m, 2H), 3.00 (s, 3H), 1.40 (t, J= 40 Hz, 3H). MS (M+H)+: 456 / 458.
Step 6 – Synthesis of 5-Bromo-2-(4-fluoro-phenyl)-6-methanesulfonylamino-benzofuran-3- carboxylic acid
To a solution of ethyl 5-bromo-2-(4-fluorophenyl)-6-(methylsulfonamido) benzofuran-3-carboxylate (53 g, 0.23 mol) in dioxane / H20 (5 / 1, 600 mL) was added
LiOH-H20 (25 g, 1.17 mol), and the mixture was stirred at 100 °C for 3 hours. After
concentrated, the resulting residue was dissolved in H20, 1 N HCl was added until pH reached 3, and the mixture was extracted with EtOAc. The organic layer was washed with brine, dried over Na2S04 and filtered. The solvent was removed to provide the product of 5-bromo-2-(4- fluorophenyl)-6-(methylsulfonamido)benzofuran-3-carboxylic acid (48 g, yield: 96%).1H- MR (DMSO- e, 400 MHz) δ 13.49 (s, 1H), 9.67 (s, 1H), 8.30 (s, 1H), 8.12-8.17 (m, 2H), 7.87 (s, 1H), 7.45-7.50 (m, 2H), 3.16 (s, 3H). MS (M+H)+: 428 / 430. Step 7 – Synthesis of 5-Bromo-2-(4-fluoro-phenyl)-6-methanesulfonylamino-benzofuran-3- carboxylic acid methylamide

A solution of 5-bromo-2-(4-fluorophenyl)-6-(methylsulfonamido) benzofuran-3- carboxylic acid (33 g, 77 mmol), HOBT (15.6 g, 115.5 mmol) and EDCI (22.2 g, 115.5 mmol) in DMF (250 mL) was stirred at room temperature. After 2 hours, Et3N (50 mL) and CH3 H2 (HC1 salt, 17.7 g, 231 mmol) was added to the mixture, and the mixture was stirred overnight. After the solvent was removed, H20 was added and the mixture was extracted with ethyl acetate. The combined organic layer was washed with H20, brine and concentrated in vacuo. The resulting residue was washed with EtOAc to provide the product of 5-bromo-2-(4-fluorophenyl)-N- methyl-6-(methylsulfonamido)benzofuran-3-carboxamide (32 g, yield: 94%). 1H- MR (DMSO- ck, 400 MHz) δ 9.55 (br s, 1H), 8.46-8.48 (m, 1H), 8.12-8.17 (m, 2H), 7.96 (s, 1H), 7.87 (s, 1H), 7.45-7.50 (m, 2H), 3.16 (s, 3H), 2.93 (d, J= 8.4 Hz, 3H). MS (M+H)+: 441 / 443.
Step 8 – Synthesis of 5-bromo-2-(4-fluorophenyl)-N-methyl-6-(N- methylmethylsulfonamido benzofuran-3-carboxamide

CH3I (31.6 g, 223 mmol) was added to a mixture of 5-bromo-2-(4-fluorophenyl)- N-methyl-6-(methylsulfonamido)benzofuran-3-carboxamide (32 g, 74 mmol), K2C03 (25.6 g, 186 mmol) and KI (246 mg, 1.5 mmol) in DMF (150 mL) under N2 protection. The mixture was stirred at 80-90 °C overnight. After concentrated in vacuo, the resulting residue was washed with water (200 mL) and EtOAc (200 mL) to provide the product of 5-bromo-2-(4- fluorophenyl)-N-methyl-6-(N-methylmethylsulfonamido)benzofuran-3-carboxamide (31.5 g, 94%). 1H- MR (CDCI3, 400 MHz) δ 8.16 (s, 1H), 7.88-7.92 (m, 2H), 7.70 (s, 1H), 7.18-7.23 (m, 2H), 5.78 (br s, 1H), 3.34 (s, 3H), 3.09 (s, 3H), 3.00 (d, J= 4.8 Hz, 3H). MS (M+H)+: 455 / 457. Step 9 – Synthesis of 2-(4-fluorophenyl)-N-methyl-6-(N-methylmethylsulfonamido)-5-(4, 4, 5, 5- tetramethyl-1 -dioxaborolan-2-yl)benzofuran-3-carboxamide
a degassed solution of 5-bromo-2-(4-fluorophenyl)-N-methyl-6-(N- methylmethylsulfonamido)benzofuran-3-carboxamide (1.0 g, 2.2 mmol) and pinacol diborane (2.79 g, 11.0 mmol) in 1,4-Dioxane (25 mL) was added KOAc (647 mg, 6.6 mmol) under N2 and stirred for 4 hours at room temperature. Then Pd(dppf)Cl2 (60 mg) was added, and the mixture was stirred for another 30 minutes. Then the mixture was put into a pre-heated oil-bath at 130 °C and stirred for another 1 hour under N2. The reaction mixture was cooled to room
temperatureand concentrated and extracted with EtOAc. The organic layers were washed with brine, dried over Na2S04. After concentrated, the crude product of the boronic ester was purified using column chromatography (petroleum ether / EtOAc = 5 / 1 to 2 / 1) to obtain 2-(4- fluorophenyl)-N-methyl-6-(N-methylmethylsulfonamido)-5-(4,4,5,5-tetramethyl-l,3,2- dioxaborolan-2-yl)benzofuran-3-carboxamide as white solid (700 mg, yield: 64%). 1H- MR (CDCI3, 400 ΜΗζ) δ 8.17 (s, 1H), 7.87-7.91 (m, 2H), 7.52 (s, 1H), 7.11 (t, 7= 7.6 Hz, 2H), 5.81 (d, 7= 2.8 Hz, 1H), 3.30 (s, 3H), 2.97 (d, 7= 5.2 Hz, 3H), 2.90 (s, 3H), 1.31 (s, 12H). MS (M+H)+: 503.
Step 10 – Synthesis of tert-butyl 4-fluoro-lH-indole-l -car boxy late
To a solution of 4-fluoro-lH-indole (5 g, 0.11 mol) and DMAP (150 mg, 3%Wt) in THF (50 mL) was added (Boc)20 (8.5 g, 0.04 mol) dropwise. The mixture was stirred at room temperature for 2 hours. The organic solvent was removed in vacuo, and the resulting residue was purified using column chromatography (pure petroleum ether) to provide tert-butyl 4-fluoro- lH-indole-l-carboxylate (8.3 g, yield: 96%). 1H- MR (CDC13, 400 MHz) δ 7.92 (d, J= 8.4 Hz, 1H), 7.55 (d, J= 3.6 Hz, 1H), 7.23 (m, 1H), 6.90 (m, 1H), 6.66 (d, J= 3.6 Hz, 1H), 1.67 (s, 9H). MS (M+H)+: 236.
Step 11 – Synthesis of (l-(tert-butoxycarbonyl)-4-fluoro-lH-indol-2-yl)boronic acid

To a solution of diisopropylamine (7.5 mL, 0.11 mol) in THF (35 mL) at 0 °C was added «-BuLi (21 mL, 0.055 mol) dropwise. The mixture was stirred at 0 °C for 40 minutes. Then the mixture was cooled to -78 °C. Tert-butyl 4-fluoro-lH-indole-l-carboxylate (5 g, 0.02 mol) in THF (13 mL) was added dropwise slowly. After addition, the mixture was stirred at -78 °C for 2 hours. Then triisopropyl borate (3.29 g, 0.03 mol) was added. The mixture was stirred at -78 °C for another 40 minutes. The reaction was monitored using TLC. When the reaction was completed, the mixture was adjusted to pH = 6 with 1 N HC1. After extracted with EtOAc (25 mL x 3), the combined organic layers were washed with brine (50 mL), dried over Na2S04, filtered and concentrated in vacuo. The obtained solid was recrystallized with EtOAc and petroleum ether to provide (l-(tert-butoxycarbonyl)-4-fluoro-lH-indol-2-yl)boronic acid (4.5 g, yield: 76.7%, which might be unstable at high temp, work up, store in fridge). 1H- MR (CDC13, 400 MHz) δ 7.77 (d, J= 8.4 Hz, 1H), 7.57 (s, 1H), 7.44 (s, 2H), 7.24 (m, 1H), 6.90 (m, 1H), 1.66 (s, 9H). MS (M+H)+: 280.
Step 12 – Synthesis of 6-chloro-2-iodopyridin-3-ol
6-chloropyridin-3-ol (5.0 g, 38.6 mmol) was dissolved in water (50 mL) and placed under an N2 atmosphere. Na2C03 (8.2 g, 77.4 mmol) was added followed by iodine (9.8 g, 38.8 mmol). The reaction mixture was stirred at room temperature for 2 hours. The mixture was poured into 1M Na2S203 and extracted with EtOAc. The combined organic phases were washed with brine, dried over Na2S04 and concentrated to provide the product of 6-chloro-2- iodopyridin-3-ol (7.0 g, yield: 70.9%). 1H- MR (CDC13, 400 MHz) δ 7.17 (d, J= 8.4 Hz, 1H), 7.06 (d, J= 8.4 Hz, 1H). MS (M+H)+: 256 / 258.
Step 13 – Synthesis of 6-chloro-2-(4-fluoro-lH-indol-2-yl)pyridin-3-ol
A mixture of (l-(tert-butoxycarbonyl)-4-fluoro-lH-indol-2-yl)boronic acid (5 g, 18.0 mmol), 6-chloro-2-iodopyridin-3-ol (3.82 g, 15.0 mol) and NaHC03 (3.78 g, 45.0 mol) in 1, 4-dioxane (76 mL) and water (7 mL) was stirred at room temperature for 15 minutes. Then Pd(PPh3)2Cl2 (527 mg, 0.75 mmol) was added under nitrogen atmosphere, and the mixture was heated at 100 °C under N2 for 16 hours. The reaction mixture was cooled to room temperature, diluted with EtOAc (50 mL), filtered and concentrated in vacuo. The resulting residue was diluted with H20 (60 mL) and EtOAc (30 mL), and the layer was separated, the aqueous layer was extracted with EtOAc (3*30 mL). The combined organic layers were washed with brine (50 mL), dried over Na2S04, filtered and concentrated in vacuo. The resulting residue was purified using column chromatography (petroleum ether / EtOAc = 20 / 1 ~ 3 / 1) to provide 6-chloro-2- (4-fluoro-lH-indol-2-yl)pyridin-3-ol (3 g, yield: 76.5%). 1H- MR (MeOD, 400 MHz) δ 7.36 (s, 1H), 7.23-7.27 (m, 2H), 7.03-7.11 (m, 2H), 6.63-6.68 (m, 1H). MS (M+H)+: 263 / 265.
Ste 14 – Synthesis of 2-chloro-ll-fluoro-6H-pyrido[2′,3′:5, 6][l,3]oxazino[3,4-a]indole
A solution of 6-chloro-2-(4-fluoro-lH-indol-2-yl)pyridin-3-ol (2 g, 7.6 mmol) and Cs2C03 (7.46 g, 22.89 mmol) in DMF (100 mL) was stirred at 100 °C (internal temperature) for 15 min, and then chloroiodomethane (2.85 g, 15.3 mmol) in DMF (2 mL) was added dropwise. After the reaction was completed, the mixture was filtered and concentrated in vacuo. The resulting residue was diluted with water (50 mL) and extracted with ethyl acetate (30 mL x 3). The organic layer was washed with brine, dried over Na2S04 and concentrated in vacuo. The resulting residue was purified using column chromatography (petroleum ether:EA=10: l) to provde 2-chloro-l l-fluoro-6H-pyrido[2′,3′:5,6][l,3]oxazino[3,4-a]indole (1.8 g, yield: 86.1%). 1H- MR (DMSO-i¾, 400 MHz) δ 7.64 (d, J= 8.8 Hz, 1H), 7.39-7.46 (m, 2H), 7.21-7.25 (m, 1H), 7.06 (s, 1H), 6.88-6.92 (m, 1H), 6.18 (s, 2H). MS (M+H)+: 275 / 277. Step 15 – Synthesis of5-(ll-fluoro-6H-pyrido[2 3′:5, 6][l,3]oxazino[3,4-a]indol-2-yl)-2-(4- fluorophenyl)-N-methyl-6-(N-methylmethylsulfonamido)benzofuran-3-carboxam
To a degassed solution of 2-(4-fluorophenyl)-N-methyl-6-(N- methylmethylsulfonamido)-5-(4,4,5,5-tetramethyl-l,3,2-dioxaborolan-2-yl)benzofuran-3- carboxamide (100 mg, 0.199 mmol), 2-chloro-l l-fluoro-6H-pyrido[2′,3′:5,6][l,3]oxazino[3,4- a]indole (56 mg, 0.199 mmol) and Κ3Ρ04·3Η20 (159 mg, 0.597 mmol) in dioxane / H20 (0.8 mL / 0.2 mL) was added Pd2(dba)3 (9 mg, 0.01 mmol) and X-Phos (9 mg, 0.02 mmol) under N2. The mixture was heated at 80 °C for 1 hour. The mixture was then diluted with water (30 mL) and extracted with EtOAc (15 mL x 3). The organic layer was washed with brine (20 mL), dried over Na2S04 and concentrated in vacuo. The resulting residue was purified using prep-TLC (petroleum ether / EtOAc = 1 : 1.5) to provde the pure product of 5-(l l-fluoro-6H- pyrido [2′, 3 ‘ : 5 , 6] [ 1 , 3 ]oxazino [3 ,4-a]indol-2-yl)-2-(4-fluorophenyl)-N-methyl-6-(N- methylmethylsulfonamido)benzofuran-3-carboxamide (60 mg, 48.8%). 1H- MR (CDC13, 400 MHz) δ: 7.99 (s, 1H), 7.93-7.96 (m, 2H), 7.65 (s, 1H), 7.45-7.50 (m, 2H), 7.17-7.21 (m, 4H), 7.10 (d, J= 8.0 Hz, 1H), 6.81-6.85 (m, 1H), 5.98 (s, 3H), 3.35 (s, 3H), 2.98 (d, J= 4.8 Hz, 3H), 2.72 (s, 3H). MS (M+H)+: 615.
Paper

We describe the route development and multikilogram-scale synthesis of an HCV NS5B site D inhibitor, MK-8876. The key topics covered are (1) process improvement of the two main fragments; (2) optimization of the initially troublesome penultimate step, a key bis(boronic acid) (BBA)-based borylation; (3) process development of the final Suzuki–Miyaura coupling; and (4) control of the drug substance form. These efforts culminated in a 28 kg delivery of the desired active pharmaceutical ingredient.
Process Development of the HCV NS5B Site D Inhibitor MK-8876
*E-mail: qinghao.chen@merck.com
http://pubs.acs.org/doi/abs/10.1021/acs.oprd.5b00405
PAPER

Using the Teasdale method, purge factor estimates for six impurities identified as mutagenic alerts in the synthesis of MK-8876 are compared to actual measured amounts of these impurities determined via appropriate analytical methods. The results from this comparison illustrate the conservative nature of purge factor estimates, meaning that overprediction of mutagenic impurity purging is unlikely when using this method. Industry and regulatory acceptance of the purge factor estimation method may help minimize analytical burden in pharmaceutical development projects.
Evaluation and Control of Mutagenic Impurities in a Development Compound: Purge Factor Estimates vs Measured Amounts
*E-mail: mark_mclaughlin@merck.com.
http://pubs.acs.org/doi/abs/10.1021/acs.oprd.5b00263?journalCode=oprdfk
| WO2004041201A2 * | Oct 31, 2003 | May 21, 2004 | Viropharma Incorporated | Benzofuran compounds, compositions and methods for treatment and prophylaxis of hepatitis c viral infections and associated diseases |
| WO2011106992A1 * | Mar 2, 2011 | Sep 9, 2011 | Merck Sharp & Dohme Corp. | Inhibitors of hepatitis c virus ns5b polymerase |
| WO2004041201A2 * | Oct 31, 2003 | May 21, 2004 | Viropharma Incorporated | Benzofuran compounds, compositions and methods for treatment and prophylaxis of hepatitis c viral infections and associated diseases |
| WO2010030592A1 * | Sep 8, 2009 | Mar 18, 2010 | Bristol-Myers Squibb Company | Compounds for the treatment of hepatitis c |
| WO2011106992A1 * | Mar 2, 2011 | Sep 9, 2011 | Merck Sharp & Dohme Corp. | Inhibitors of hepatitis c virus ns5b polymerase |
| Citing Patent | Filing date | Publication date | Applicant | Title |
|---|---|---|---|---|
| WO2014123794A1 * | Feb 3, 2014 | Aug 14, 2014 | Merck Sharp & Dohme Corp. | Tetracyclic heterocycle compounds and methods of use thereof for the treatment of hepatitis c |
| WO2014123795A2 * | Feb 3, 2014 | Aug 14, 2014 | Merck Sharp & Dohme Corp. | Tetracyclic heterocycle compounds and methods of use thereof for the treatment of hepatitis c |
| WO2014123795A3 * | Feb 3, 2014 | Oct 30, 2014 | Merck Sharp & Dohme Corp. | Tetracyclic heterocycle compounds and methods of use thereof for the treatment of hepatitis c |
| US9242998 | Feb 3, 2014 | Jan 26, 2016 | Merck Sharp & Dohme Corp. | Tetracyclic heterocycle compounds and methods of use thereof for the treatment of hepatitis C |
//////MK-8876, 1426960-33-9, Merck & Co, Antivirals, Phase I, Hepatitis C
Fc7cccc6c7cc2n6COc1ccc(nc12)c3cc4c(cc3N(C)S(C)(=O)=O)oc(c4C(=O)NC)c5ccc(F)cc5
BRISTOL-MYERS SQUIBB’S TRICYCLOHEXADECAHEXAENE DERIVATIVES FOR USE IN THE TREATMENT OF HEPATITIS C VIRUS
 .
TRICYCLOHEXADECAHEXAENE DERIVATIVES FOR USE IN THE TREATMENT OF HEPATITIS C VIRUS
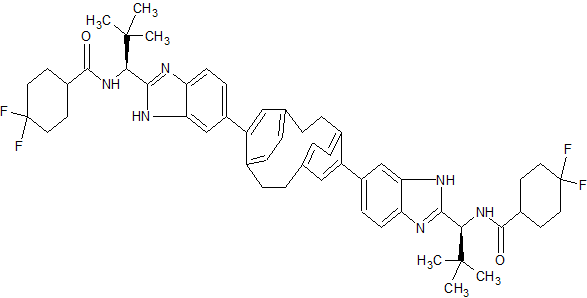
WO2015026454, COMBINATIONS COMPRISING TRICYCLOHEXADECAHEXAENE DERIVATIVES FOR USE IN THE TREATMENT OF HEPATITIS C VIRUS
BRISTOL-MYERS SQUIBB COMPANY [US/US]; Route 206 and Province Line Road Princeton, New Jersey 08543 (US)
PATENT WO2015026454 [LINK]
WANG, Alan Xiangdong; (US).
LOPEZ, Omar D.; (US).
TU, Yong; (US).
BELEMA, Makonen; (US)
Example B-l
Example B-l Step a

To a solution of 4-bromobenzene-l,2-diamine (2.5 g, 13.37 mmol) in DCM (30 mL) was added (S)-2-((tert-butoxycarbonyl)amino)-3,3-dimethylbutanoic acid (3.09 g, 13.37 mmol), DIPEA (2.334 mL, 13.37 mmol) and HATU (5.08 g, 13.37 mmol). The reaction mixture was stirred at room temperature for 18 h. The reaction mixture was diluted with water and extracted with DCM. The organic phase was washed with brine, dried over Na2S04, filtered and concentrated. The crude material was purified by ISCO using 40 g Redisep silica column, CHCl3/MeOH as eluant to obtain (S)-tert-butyl ( 1 -((2-amino-4-bromophenyl)amino)-3 ,3 -dimethyl- 1 -oxobutan-2-yl) carbamate (1.82 g) as yellow solid. LC (Condition 1): Rt = 2.13 min. LC/MS: Anal. Calcd. for [M+H20]+ Ci7H27BrN204 : 402.12; found 402.2. 1H NMR (DMSO-d6, δ = 2.50 ppm, 400 MHz): δ 9.35 – 9.21 (m, 1 H), 7.07 (d, J= 8.5 Hz, 1 H), 6.91 (d, J= 2.0 Hz, 1 H), 6.80 – 6.60 (m, 1 H), 5.25 – 5.01 (m, 2 H), 4.07 – 3.89 (m, 1 H), 1.52 – 1.34 (m, 9 H), 1.02 – 0.86 (m, 9 H).
Example B-l, Step b

Acetic acid (15 mL) was added to (S)-tert-butyl (l-((2-amino-4-bromo phenyl)amino)-3,3-dimethyl-l-oxobutan-2-yl)carbamate (1.8 g, 4.50 mmol) and the reaction mixture was heated to 65 °C for overnight. The volatile component was removed in vacuo, and the residue was co-evaporated with dry CH2C12 (2 x 15 mL). The organic phase was washed with saturated NaHC03 solution, brine, dried over Na2S04 and concentrated to obtain (S)-tert-butyl (l-(6-bromo-lH-benzo[d] imidazol-2-yl)-2,2-dimethyl propyl)carbamate (1.68 g) as yellow solid. LC (Condition 1): Rt = 2.19 min. LC/MS: Anal. Calcd. for [M+H]+ Ci7H25BrN302 : 381.11; found 382.2. 1H NMR (DMSO-dg, δ = 2.50 ppm, 300 MHz): δ 12.46 – 12.27 (m, 1 H), 7.82 – 7.65 (m, 1 H), 7.59 – 7.41 (m, 1 H), 7.29 (dt, J= 1.9, 8.5 Hz, 1 H), 7.12 – 6.90 (m, 1 H), 4.64 (d, J= 9.8 Hz, 1 H), 1.44 – 1.27 (m, 9 H), 0.88 (br. s., 9 H).
-1 Step c

To a solution of (S)-tert-butyl (l-(6-bromo-lH-benzo[d]imidazol-2-yl)-2,2-dimethyl propyl) carbamate (1.57 g, 4.11 mmol) in dioxane (25 mL) was added bis (pinacolato)diboron (1.564 g, 6.16 mmol) and potassium acetate (1.209 g, 12.32 mmol). The reaction mixture was purged with argon for 10 min then PdCl2(dppf) (0.150 g, 0.205 mmol) was added to the above reaction mixture and again purged with argon for 5 min. The reaction mixture was heated to 90 °C for overnight. The reaction mixture was diluted with water (15 ml) and extracted with EtOAc (2 x 25 ml). The combined organic phase was washed with brine, dried over Na2S04 and concentrated in vacuo. The crude material was purified by ISCO using 40 g Redisep column, hexane/ethyl acetate as eluant to afford (S)-tert-butyl (2,2-dimethyl-l-(6-(4,4,5 ,5-tetramethyl- 1 ,3 ,2-dioxaborolan-2-yl)- 1 H-benzo[d]imidazol-2-yl)propyl) carbamate (1.35 g) as yellow solid. LC (Condition 1): Rt = 2.21 min. LC/MS: Anal. Calcd. for [M+H]+ C23H37BN304 : 430.29; found 430.4. 1H NMR (CD3OD, δ = 3.34 ppm, 400 MHz): δ 7.98 (s, 1 H), 7.65 (dd, J= 1.0, 8.5 Hz, 1 H), 7.53(d, J= 8.5 Hz, 1 H), 4.73 (br. s., 1 H), 1.37 (s, 12 H), 1.24 (m, 9 H), 1.01 (s, 9 H).
-1 Step d

To a solution of (S)-tert-butyl (2,2-dimethyl-l-(6-(4,4,5,5-tetramethyl-l,3,2-dioxaborolan-2-yl)-lH-benzo[d]imidazol-2-yl)propyl)carbamate (1.114 g, 2.59 mmol) and 4,16-dibromo[2,2]paracyclophane (0.38g, 1.038 mmol) in dioxane (10 mL) was added Cs2C03 (0.845 g, 2.59 mmol) in water (2 mL) and degassed for 10 min.
PdCl2(dppf) (0.038 g, 0.052 mmol) was added to the above reaction mixture and again degassed for 5 min. The reaction mixture was heated to 90 °C for 12 h. Then the reaction mixture was filtered to get Example B-1 Step d which was taken for next step without further purification. LC (Condition 1): Rt = 2.54 min. LC/MS: Anal. Calcd. for [M+H]+ ^0Η63Ν6Ο4 : 811.49; found 811.6. 1H NMR (DMSO-d6, δ = 2.50 ppm, 300 MHz): δ 12.36 (br. s., 2 H), 7.85 – 7.52 (m, 4 H), 7.32 (d, J= 7.9 Hz, 2 H), 7.05 (br. s., 2 H), 6.89 – 6.67 (m, 4 H), 6.54 (br. s., 2 H), 4.72 (d, J= 8.7 Hz, 2 H), 3.57 – 3.44 (m, 2 H), 3.07 (br. s., 2 H), 2.83 (br. s., 2 H), 2.65 (br. s., 2 H), 1.36 (s, 18 H), 1.08 – 0.91 (m, 18 H).
-1 Step e

HC1 in dioxane (4 mL, 24.00 mmol) was added to Example B-1 Step d (0.1 g,
0.102 mmol), and the reaction mixture was allowed to stir at RT for 2 h. Completion of the reaction was monitored by LCMS. The volatile component was removed in vacuo and the residue was washed with diethyl ether and dried to afford Example B-1 Step e (0.07 g) as yellow solid. LC (Condition 1): R, = 2.54 min. LC/MS: Anal.
Calcd. for [M+H]+ C40H47N6 : 611.39; found 611.4. 1H NMR (CD3OD, δ = 3.34 ppm, 400 MHz): δ 7.90 (d, J= 13.1 Hz, 2 H), 7.83 (d, J= 8.5 Hz, 2 H), 7.61 (d, J= 8.5 Hz, 2 H), 6.84 (d, J= 6.5 Hz, 2 H), 6.78 (s, 2 H), 6.70 – 6.65 (m, 2 H), 4.54 (d, J= 1.0 Hz, 2 H), 3.54 – 3.46 (m, 2 H), 3.18 – 3.10 (m, 2 H), 2.98 – 2.86 (m, 2 H), 2.71 (br. s., 2 H), 1.25 – 1.22 (m, 18 H).
To a solution of Example B-1 Step e (0.04 g, 0.053 mmol) in DMF (5 mL) was added 4,4-difluorocyclohexanecarboxylic acid (0.017 g, 0.106 mmol), DIPEA (0.055 mL, 0.317 mmol) and HATU (0.030 g, 0.079 mmol). After being stirred for 2 h at room temperature, the volatile component was removed in vacuo and the residue was dissolved in DCM (10 mL), washed with saturated solution of NH4C1, 10% NaHC03 solution, brine, dried over Na2S04 and concentrated in vacuo. The crude was purified by reverse phase HPLC purification to give Example B-1 as a white solid. LC (Condition 1): R, = 2.37 min. LC/MS: Anal. Calcd. for [M+H]+
C54H63F4N602: 903.49; found 903.4. 1H NMR (DMSO-d6, δ = 2.50 ppm, 400 MHz): δ 12.53 – 12.32 (m, 2 H), 8.41 – 8.21 (m, 2 H), 7.84 – 7.50 (m, 4 H), 7.43 – 7.24 (m, 2 H), 6.90 – 6.67 (m, 4 H), 6.60 – 6.44 (m, 2 H), 5.14 – 4.97 (m, 2 H), 3.44 (br. s., 2 H), 3.08 (br. s., 2 H), 2.93 – 2.77 (m, 2 H), 2.73 – 2.56 (m, 4 H), 2.20 – 1.98 (m, 3 H), 1.96 – 1.49 (m, 13 H), 1.02 (s, 18 H).
Starting materials can be obtained from commercial sources or prepared by well-established literature methods known to those of ordinary skill in the art. Acid precursors for the final step can be prepared according to the methods described in U.S. Patent Application Serial No. 13/933495, filed July 2, 2013.
LC/MS Condition 1
Column = Ascentis Express C18, 2.1 X 50 mm, 2.7 um
Solvent A = CH3CN (2%) + 10 mM NH4COOH in H20 (98%)
Solvent B = CH3CN (98%) + 10 mM NH4COOH in H20 (2%)
Start %B = 0; Final %B = 100
Gradient time = 1.4 min; Stop time = 4 min
Stop time = 4 min
Flow Rate = 1 mL/min; Wavelength = 220 nm
LC/MS Condition 2
Column = Waters BEH CI 8, 2.0 x 50 mm, 1.7 μιη
Slovent A = ACN (5%) + H20 (95%) containing 10 mM NH4OAc
Solvent B = ACN (95%) + H20 (5%) containing 10 mM NH4OAc
Start %B = 0; Final %B = 100
Gradient time = 3 min
Flow Rate = 1 mL/min
Wavelength = 220 nm
Temperature = 50 °C
LC/MS Condition 3
Column: Waters Phenomenex CI 8, 2.0 x 30 mm, 3 μιη particle
Mobile Phase A: 10% MeOH:90% Water :0.1%TFA
Mobile Phase B: 90% MeOH: 10% Water :0.1%TFA
Gradient: 0%B, 0-100% B over 3 minutes, then a 1 -minute hold at 100% B Flow: 0.8mL/min
Detection: 220 nm
Temperature: 40 °C
LC/MS Condition 4
Column: Waters BEH CI 8, 2.0 x 50 mm, 1.7 μιη particle
Mobile Phase A: 5:95 acetonitrile: water with 10 mM ammonium acetate Mobile Phase B: 95:5 acetonitrile: water with 10 mM ammonium acetate Gradient: 0%B, 0-100% B over 3 minutes, then a 0.5-minute hold at 100% B Flow: 1 mL/min
Detection: UV at 220 nm
Temperature: 50 °C
/////////1663477-91-5, BRISTOL-MYERS SQUIBB, TRICYCLOHEXADECAHEXAENE DERIVATIVES, TREATMENT OF HEPATITIS C VIRUS
FC1(F)CCC(CC1)C(=O)N[C@H](c2nc3ccc(cc3n2)c9cc4ccc9CCc5ccc(CC4)c(c5)c6ccc7nc(nc7c6)[C@@H](NC(=O)C8CCC(F)(F)CC8)C(C)(C)C)C(C)(C)C
Ripasudil hydrochloride hydrate 塩酸塩水和物 , リパスジル
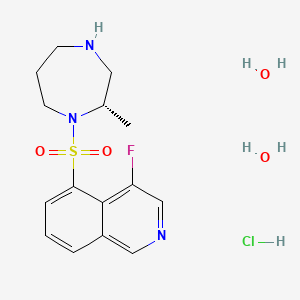
Ripasudil hydrochloride hydrate
4-fluoro-5-[[(2S)-2-methyl-1,4-diazepan-1-yl]sulfonyl]isoquinoline;dihydrate;hydrochloride
4-Fluoro-5-[2(S)-methylperhydro-1,4-diazepin-1-ylsulfonyl]isoquinoline hydrochloride dihydrate
(S)-4-Fluoro-5-(2-methyl-1,4-diazepan-1-ylsulfonyl)isoquinoline hydrochloride dihydrate
Cas 223645-67-8 FREE
| M.Wt | 395.88 | OR C15H18FN3O2S·HCl·2H2O | |
|---|---|---|---|
| Formula | C15H23ClFN3O4S | ||
| CAS No | 887375-67-9 .HCL 2 H2O |
016TTR32QF, K 115
LAUNCHED 2014 Kowa
JAPAN 2014-09-26, Glanatec

リパスジル塩酸塩水和物
Ripasudil Hydrochloride Hydrate

C15H18FN3O2S.HCl.2H2O : 395.88
[887375-67-9]
SEE http://pdf.irpocket.com/C4576/GpH7/tLM4/sJIT.pdf
Ripasudil
- Molecular FormulaC15H18FN3O2S
- Average mass323.386
| Company | D. Western Therapeutics Institute Inc. |
| Description | Selective rho kinase inhibitor |
| Molecular Target | Rho kinase |
| Mechanism of Action | Rho kinase inhibitor |
SEE
NMR ETC
Published online: 24th May, 2011
■ A Practical Synthesis of Novel Rho-Kinase Inhibitor, (S)-4-Fluoro-5-(2-methyl-1,4-diazepan-1-ylsulfonyl)isoquinoline
Noriaki Gomi, Tadaaki Ohgiya, Kimiyuki Shibuya,* Jyunji Katsuyama, Masayuki Masumoto, and Hitoshi Sakai
*Pharmaceutical Division, Tokyo New Drug Research Laboratories, Kowa Co., Ltd., 2-17-43, Noguchicho, Higashimurayama, Tokyo 189-0022, Japan
Abstract
A practical synthesis of novel Rho-kinase inhibitor, (S)-4-fluoro-5-(2-methyl-1,4-diazepan-1-ylsulfonyl)isoquinoline hydrochloride dihydrate (1) was achieved in a pilot-scale production. We have demonstrated the regioselective chlorosulfonylation of 4-fluoroisoquinoline in an one-pot reaction to afford 4-fluoroisoquinoline-5-sulfonyl chloride and the asymmetric construction of the (S)-2-methyl-1,4-diazepane moiety as key steps.
K-115, an isoquinolinesulfonamide compound, is a highly selective and potent (IC50 = 31 nM) Rho-kinase inhibitor; is in Phase II clinical development in patients with POAG or ocular hypertension.Ripasudil hydrochloride hydrate (Glanatec® ophthalmic solution 0.4 %; hereafter referred to as ripasudil) is a small-molecule, Rho-associated kinase inhibitor developed by Kowa Company, Ltd. for the treatment of glaucoma and ocular hypertension. This compound, which was originally discovered by D. Western Therapeutics Institute, Inc., reduces intraocular pressure (IOP) by directly acting on the trabecular meshwork, thereby increasing conventional outflow through the Schlemm’s canal.
Ripasudil hydrochloride hydrate was first approved by Pharmaceuticals and Medical Devices Agency of Japan (PMDA) on Sept 26, 2014. It was developed and marketed as Glanatek® by Kowa Pharmaceuticals.
Ripasudil hydrochloride hydrate is the first drug that can inhibit the rho-associated, coiled-coil containing protein kinase (ROCK). It is indicated for the treatment of glaucoma and ocular hypertension.
Glanatek® is available as solution (0.4%) for ophthalmic use, containing 4 mg of free Ripasudil per millimeter, and the recommended dose is one drop twice daily.
As a result of this mechanism of action, ripasudil may offer additive effects in the treatment of glaucoma and ocular hypertension when used in combination with agents such as prostaglandin analogues (which increase uveoscleral outflow) and β blockers (which reduce aqueous production).
The eye drop product has been approved in Japan for the twice-daily treatment of glaucoma and ocular hypertension, when other therapeutic agents are not effective or cannot be administered. Phase II study is underway for the treatment of diabetic retinopathy.
K-115 is a Rho-kinase inhibitor as ophthalmic solution originally developed by Kowa and D Western Therapeutics Institute (DWTI). The product candidate was approved and launched in Japan for the treatment of glaucoma and ocular hypertension in 2014.
In 2002, the compound was licensed to Kowa Pharmaceutical by D Western Therapeutics Institute (DWTI) in Japan for the treatment of glaucoma. The compound is currently in phase II clinical trials at the company for the treatment of age-related macular degeneration and diabetic retinopathy.
Use of (S)-(-)-1-(4- fluoro-5-isoquinoline-sulfonyl)-2-methyl-1,4-homopiperazine (ripasudil hydrochloride, first disclosed in WO9920620), in the form of eye drops, for the treatment of retinal diseases, particularly diabetic retinopathy or age-related macular degeneration.
Follows on from WO2012105674 by claiming a combination of the same compound. Kowa, under license from D Western Therapeutics Institute, has developed the Rho kinase inhibitor ripasudil hydrochloride hydrate (presumed to be Glanatek) as an eye drop formulation for the treatment of glaucoma and ocular hypertension which was approved in Japan in September 2014..
The company is also developing the agent for the treatment of diabetic retinopathy, for which it is in phase II trial as of October 2014.
Ripasudil (Glanatec) is a drug used for the treatment of glaucoma and ocular hypertension. It is approved for use in Japan as a 0.4% ophthalmic solution.[1]
Ripasudil, a derivative of fasudil, is a rho kinase inhibitor.[2]
Paper
A Practical Synthesis of (S)-tert-butyl 3-methyl-1,4-diazepane-1-carboxylate, the key intermediate of Rho-kinase inhibitor K-115
Synthesis (Stuttgart) 2012, 44(20): 3171
https://www.thieme-connect.com/products/ejournals/abstract/10.1055/s-0032-1316771
practical synthesis of (S)-tert-butyl 3-methyl-1,4-diazepane-1-carboxylate has been established for supplying this key intermediate of Rho–kinase inhibitor K-115 in a multikilogram production. The chiral 1,4-diazepane was constructed by intramolecular Fukuyama–Mitsunobu cyclization of a N-nosyl diamino alcohol starting from the commercially available (S)- or (R)-2-aminopropan-1-ol. In the same manner, an enantiomeric pair of a structural isomer were prepared for demonstration of the synthetic utility.
SEE
WO 2006137368 http://www.google.com/patents/WO2006137368A1?cl=en
PATENT
WO 2012026529
http://www.google.com/patents/WO2012026529A1?cl=en
The including prevention and treatment cerebral infarction, cerebral hemorrhage, subarachnoid hemorrhage, cerebrovascular disorders such as cerebral edema, the present invention relates to a salt thereof or isoquinoline derivatives useful as therapeutic agents, particularly glaucoma.
(S) – (-) -1 – (4 – fluoro-iso-5 – yl) sulfonyl – 2 – methyl -1,4 – diazepane the following formula (1):

It is a compound represented by the particular it is a crystalline water-soluble, not hygroscopic, because it is excellent in chemical stability, it is useful as a medicament has been known for its hydrochloride dihydrate ( refer to Patent Documents 1 and 2). -5 Isoquinoline of these – the sulfonamide compounds, that prophylactic and therapeutic agents for cerebral infarction, cerebral hemorrhage, subarachnoid hemorrhage, cerebrovascular disorders such as cerebral edema, is useful as a therapeutic agent for preventing and glaucoma in particular is known (1-5 see Patent Document 1).
Conventionally, for example, a method of manufacturing by the method described in Patent Document 1, as shown in the following production process has been reported preparation of said compound (Production Method 1-A).

That is, (S)-1-tert-butoxycarbonyl – 3 – by reacting the presence of triethylamine in methylene chloride-fluoro-isoquinoline (2) – methyl -1,4 – diazepane and 5 (3) – chloro-sulfonyl -4 by adding trifluoroacetic acid in methylene chloride compound (the first step), obtained following (4) to synthesize a compound (4) by deprotection to (second step) the desired compound (1) This is a method of manufacturing.
It is also an important intermediate for preparing the compound (1) (S)-1-tert-butoxycarbonyl – 3 – methyl-1 ,4 – diazepane to (3), for example, in the following manner (; see JP Production Process 1-B) that can be produced is known.

Further, on the other hand, the compound (1) (see Patent Document 1) to be manufactured manufacturing routes such as: Any (Process 2) are known.

WO 1999/20620 pamphlet WO 2006/057397 pamphlet WO 1997/028130 pamphlet JP Patent Publication No. 2006-348028 JP Patent Publication No. 2006-290827
However, it is possible to produce in the laboratory of a small amount scale, but you place the point of view for mass industrial production, environmentally harmful halogenated hydrocarbon solvent in the compound of the above-mentioned process for producing 1-A is ( problem because it is carried out coupling step (3) and 2), giving significant adverse environmental exists. Therefore, solvent of halogenated hydrocarbon other than those listed to the specification of the patent document 1, for example, I tried actually dioxane, tetrahydrofuran and the like, but the present coupling reaction will be some progress indeed, Problems reaction is not completed raw material remained even after prolonged reaction time, yield undesirably stays in at most 30% was found. Furthermore, it is hard to decompose in the environment, elimination is also difficult to dioxane is not preferred irritating to humans, and are known as compounds that potentially harmful brain, kidney and liver .
When we actually produced compound (3) by the above production method 1-B, can be obtained desired compound in good yield merged with reproducibility is difficult has further been found that. That is, in the production path, 1,4 – and is used sodium hydride with dimethyl sulfoxide in forming a diazepane ring, except that I actually doing this step, Tsu than the reproducibility of the desired compound It could not be obtained in high yield Te. Also, that this is due to the synthetic route through the unstable intermediate, that it would be converted into another compound easily found this way. limitations and potential problems of the present production process is exposed since this stability may affect the reproducibility of the reaction.
Meanwhile, an attempt to carry out mass production is actually in the Process 2, it encounters various problems. For example, it is stored as an impurity whenever I repeat step, by-products formed in each stage by tandem production process ranging from step 8 gave more complex impurity profile. Depending, it is necessary to repeat a complicated recrystallization purity obtained as a medicine until the purification, the yield in the laboratory be a good overall yield is significantly reduced in the mass production of actual example be away, it does not have industrial utility of true was found. It can be summarized as follows: Considering from the viewpoint of GMP process control required for pharmaceutical production these problems.
Requires control process and numerous complex ranging 1) to 8 step, 3 2) third step – amino-1 – in the step of reacting a propanol, a difficult to remove positional isomers are mixed, 3) The fourth step water is mixed by the minute liquid extraction operation at the time of return to the free base from oxalate require crystallization purification by oxalate in the removal of contaminants of positional isomers, in 4) fifth step, 5) sixth step The Mitsunobu by reproducibility poor require water control in the Mitsunobu reaction used in the ring closure compounds to (1) compounds in (6), 6) ring closure reaction, departing management of the reagent added or the like is generated, in 7) Seventh Step it takes a complicated purification in impurity removal after the reaction, resulting in a decrease in isolated yield. These are issues that must be solved in order to provide a stable supply of raw material for pharmaceuticals high chemical purity is required.
Thus, gentle salt thereof, or the environment isoquinoline derivative comprising a compound represented by the formula (1), the present invention provides a novel production method having good reproducibility and high purity easily and in high yield I intended.
As a result of intensive studies in view of such circumstances, the present inventors, in the manufacturing process of the final target compound shown by the following expression

(Wherein represents a fluorine, chlorine, bromine or iodine, may, R 3 and 1, R 2 R represents a C 1-4 alkyl group be the same or different from each other, and P, X 1 is a protecting group shows a, 0 to m represents an integer of 3, 0 to n is. represents an integer of 3)
Is a urea-based solvents nitrile solvents, amide solvents, sulfoxide or solvents, the solvent may be preferably used in the coupling step of the compound (III) and (II) are generally very short time With these solvents It has been found that can be converted to the desired product quantitatively. It is possible to carry out the coupling step Volume scale while maintaining a high yield by using these solvents, there is no need to use a halogenated hydrocarbon solvent to give significant adverse environment. In consideration of the process such as removal of the solvent after the reaction was further found that acetonitrile is the best among these solvents. Also, since by using hydrochloric acid with ethyl acetate solvent in step deprotection can be isolated as crystal of hydrochloride desired compound (I), without going through the manipulation of solvent evaporation complicated , it has been found that it is possible to obtain the object compound (I) is a simpler operating procedure. Since there is no need to use a halogenated hydrocarbon solvent in this deprotection step further, there is no possibility of harming the environment.
It has been found that it is possible in mass production of (II), leading to the target compound purity, in high yield with good reproducibility as compared with the conventional method compounds are important intermediates in the coupling step further. That is, was it possible to lead to the intermediate high purity and in high yield by eliminating the production of a harmful halogenated hydrocarbon solvent to the environment in this manner. 1,4 addition – in order to avoid the problems encountered in the reaction using sodium hydride in dimethyl sulfoxide in forming the diazepane ring, in order to allow the cyclization reaction at mild conditions more, as a protecting group By performing the Mitsunobu reaction using Noshiru group instead of the carbobenzyloxy group, in addition to one step shorten the manufacturing process of the whole, without deteriorating the optical purity was successfully obtained the desired compound desired.

SEE

CLIP
Ripasudil hydrochloride hydrate (Glanatec)
Ripasudil hydrochloride hydrate (Glanatec) was approved in Japan in 2014 for the treatment of glaucoma and ocular hypertension.
219 Originally discovered by D. Western Therapeutics Institute,Inc. and licensed by the Kowa Company, Ltd, ripasudil
functions as a selective Rho-kinase inhibitor and reduces intraocular pressure by stimulation of aqueous humour drainage of the
trabecular meshwork.219–221
While this recent approval allows for use of ripasudil as a twice-daily monotherapy treatment when
other drugs cannot be used or are not effective, clinical trials using ripasudil as a combination therapy with other glaucoma
drugs have shown promising results in the treatment of primary open-angle glaucoma or ocular hypertension.222,223 Currently, the
Kowa Company is also pursuing trials focused on the use of ripasudil for the treatment of diabetic retinopathy and diabetic macular edema.224
While initial synthetic routes to ripasudil were carried out via a stepwise functionalization of 4-fluoroisoquinoline-5-sulfonylchloride (238),225,226 more recent reports describe an efficient route to ripasudil employing a late stage-coupling of Boc-diazepane
(237) with 4-fluoroisoquinoline-5-sulfonyl chloride (238), enabling synthesis on multi-kilogram scale and isolation of the
drug in high purity (Scheme 40).221,227,228 This optimized route to ripasudil begins with 2-nitrobenzene sulfonyl chloride (NsCl)-
mediated protection of (S)-2-amino-1-propanol (234) in 82% yield.
In this case, use of the NaHCO3/THF/H2O conditions were essential for preventing bis-nosylation.228 Alcohol activation with methanesulfonyl chloride (MsCl) in N-methyl morpholine (NMM) took place smoothly to give the corresponding mesylate 235 in 91%
yield. Direct mesylate displacement with 3-aminopropanol and subsequent amine protection as the carbamate ((Boc)2O) in a
one-pot fashion provided the corresponding Boc-amino propanol product 236 in 95% yield over 2 steps.
With the acyclic diazepane precursor 236 in hand, employment of the intramolecular Fukuyama-Mitsunobu N-alkyl cyclization conditions (diisopropylazodicarboxylate (DIAD)/PPh3) allowed generation of the diazepane in 75% yield. Nosyl group cleavage with thiophenol/K2CO3provided the Boc-diazepane 237 in 65% overall yield and 98% purity following a pH-controlled aqueous workup.
Finally, 4-fluoroisoquinoline- 5-sulfonyl chloride (238)—prepared via subjection of 4- fluoroisoquinoline (239, Scheme 41)229 to sulfur trioxide and sulfuric acid followed by treatment with thionyl chloride and finally 4 N HCl in ethyl acetate—was involved in a 1-pot, two-step procedure in which this sulfonyl chloride was coupled with diazepane 237 (TEA/MeCN) to access the ripasudil framework in quantitative yield.
Synthesis of the final drug target by deprotection with 4 MHCl in ethyl acetate followed by neutralization with aqueoussodium hydroxide provided the free base of ripasudil in 93% yield and 99.8% purity. Conversion to the more stable hydrochloride dihydrate form could be performed by treatment of the free base with 1 M HCl/EtOH and subsequent heating of the hydrochloride in H2O/acetone to provide ripasudil hydrochloride dihydrate XXIX in 83% yield.230,231
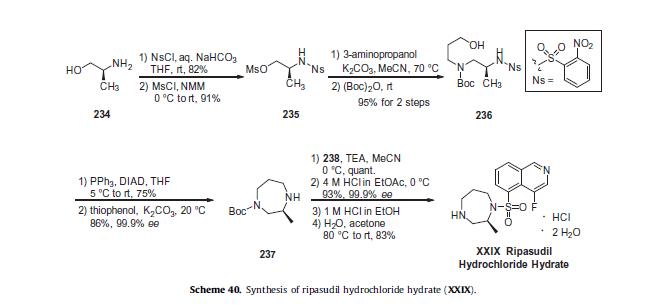
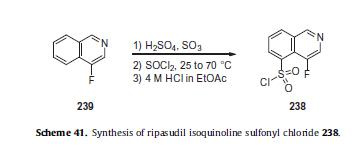
219. Garnock, J. P. K. Drugs 2014, 74, 2211.
220. Isobe, T.; Mizuno, K.; Kaneko, Y.; Ohta, M.; Koide, T.; Tanabe, S. Curr. Eye Res.2014, 39, 813.
221. Sumi, K.; Inoue, Y.; Nishio, M.; Naito, Y.; Hosoya, T.; Suzuki, M.; Hidaka, H.
Bioorg. Med. Chem. Lett. 2014, 24, 831.
222. Mizuno, K. WO Patent 2,012,105,674, 2012.
223. Mizuno, K.; Matsumoto, J. WO Patent 2,007,007,737, 2007.
224. http://clinicaltrials.jp/user/cteDetail.jsp.
225. Gomi, N.; Ohgiya, T.; Shibuya, K. WO Patent 2,012,026,529, 2012.
226. Hidaka, H.; Nishio, M.; Sumi, K. US Patent 20,080,064,681, 2008.
227. Gomi, N.; Kouketsu, A.; Ohgiya, T.; Shibuya, K. Synthesis 2012, 44, 3171.
228. Gomi, N.; Ohgiya, T.; Shibuya, K.; Katsuyama, J.; Masumoto, M.; Sakai, H.Heterocycles 2011, 83, 1771.
229. Sakai, H.; Masunoto, M.; Katsuyama, J.; Onogi, K. WO Patent 2006090783A1,2006.
230. Hidaka, H.; Matsuura, A. WO Patent 1999020620A1, 1999.
231. Ohshima, T.; Hidaka, H.; Shiratsuchi, M.; Onogi, K.; Oda, T. US Patent7858615B2, 2008.
 CAS NO. 223645-67-8, 4-fluoro-5-[[(2S)-2-methyl-1,4-diazepan-1-yl]sulfonyl]isoquinoline H-NMR spectral analysis |
 CAS NO. 223645-67-8, 4-fluoro-5-[[(2S)-2-methyl-1,4-diazepan-1-yl]sulfonyl]isoquinoline C-NMR spectral analysis |
| WO1997028130A1 | Jan 31, 1997 | Aug 7, 1997 | Hiroyoshi Hidaka | Isoquinoline derivatives and drugs |
| WO1999020620A1 | Oct 22, 1998 | Apr 29, 1999 | Hiroyoshi Hidaka | Isoquinoline derivative and drug |
| WO2006057397A1 | Nov 29, 2005 | Jun 1, 2006 | Hiroyoshi Hidaka | (s)-(-)-1-(4-fluoroisoquinolin-5-yl)sulfonyl-2-methyl-1,4homopiperazine hydrochloride dihydrate |
| JP2006290827A | Title not available | |||
| JP2006348028A | Title not available | |||
| JPH11171885A * | Title not available | |||
| JPS61227581A * | Title not available |
References
- Garnock-Jones, K. P. (2014). “Ripasudil: First global approval”. Drugs 74 (18): 2211–5. doi:10.1007/s40265-014-0333-2.PMID 25414122.
- Tanihara, H; Inoue, T; Yamamoto, T; Kuwayama, Y; Abe, H; Suganami, H; Araie, M; the K-115 Clinical Study Group (2014). “Intra-ocular pressure-lowering effects of a Rho kinase inhibitor, ripasudil (K-115), over 24 hours in primary open-angle glaucoma and ocular hypertension: A randomized, open-label, crossover study”. Acta Ophthalmologica: n/a. doi:10.1111/aos.12599. PMID 25487877.
 |
|
| Systematic (IUPAC) name | |
|---|---|
|
4-Fluoro-5-{[(2S)-2-methyl-1,4-diazepan-1-yl]sulfonyl}isoquinoline
|
|
| Clinical data | |
| Trade names | Glanatec |
| Identifiers | |
| PubChem | CID 9863672 |
| ChemSpider | 8039366 |
| Synonyms | K-115 |
| Chemical data | |
| Formula | C15H18FN3O2S |
| Molar mass | 323.39 g/mol |
///////////////// , Ripasudil hydrochloride hydrate, Ripasudil, 223645-67-8, 塩酸塩水和物 , リパスジル
O=S(=O)(c2c1c(F)cncc1ccc2)N3[C@H](CNCCC3)C
Annex 16: How a QP should handle unexpected Deviations

In a recent blog of the MHRA, the inspectorate looks at one aspect of the new Annex 16 – the handling of unexpected deviations.
In a recent blog of the U.K. Medicines and Healthcare products Regulatory Agency (MHRA), the inspectorate looks at one aspect of the new Annex 16 – the handling of unexpected deviations.
Before Annex 16 was revised, the handling of minor deviations from defined processes was discussed in the European Medicines Agency’s “reflection paper” EMEA/INS/GMP/227075/2008. However, the status of this paper was not always clear, and its use was not consistently applied. Now section 3 of the new Annex 16 provides guidance on when a Qualified Person (QP) may consider confirming compliance or certifying a batch where an unexpected deviation (concerning the manufacturing process and/or the analytical control methods) from the MA and/or GMP has occurred.
Pre-requisites
Before a QP releases a batch these pre-requisites…
View original post 226 more words
FDA publishes Technical Guide on Quality Metrics

The FDA has published a supplementing Guide on Quality Metrics. This is a very unusual step as the contents of the guide are planned to be integrated into the Guideline on Quality Metrics which hasn’t been finalised yet. Read more about the Technical Quality Metrics Guide.
The FDA has published a supplementing Guide on Quality Metrics. This is a very unusual step as the contents of the guide are planned to be integrated into the Guideline on Quality Metrics which hasn’t been finalised yet.
The so-called FDA Quality Metrics Technical Conformance Guide should supplement the Guidance for Industry: Request for Quality Metrics published on 28 July 2015 which is currently still in the draft version. We have recently published a GMP News about a Quality Metrics Case Study at Aenovaregarding a possible implementation. Now, the Technical Guide defines how the industry should submit Quality Metrics to the…
View original post 218 more words
Yonkenafil

Yonkenafil
Mw 487.61, MF C₂₄H₃₃N₅O₄S,
Cas 804518-63-6
4H-Pyrrolo[2,3-d]pyrimidin-4-one, 2-[2-ethoxy-5-[(4-ethyl-1-piperazinyl)sulfonyl]phenyl]-3,7-dihydro-5-methyl-7-propyl-,
2- [2-ethoxy –5- (4 – ethylpiperazine -1– sulfonyl) phenyl] -5 – methyl – 7 – n-Propyl-3 7 – PYRROLINE [2, 3 – d] pyrimidin – 4 – one
Phase2 Erectile dysfunction
扬子江药业 (Originator), 天士力制药 (Originator)
phosphodiesterase type 5 (PDE5) inhibitor
- Originator Tasly Pharmaceutical Group; Yangtze River Pharmaceutical Group
- Class Erectile dysfunction therapies
- Mechanism of Action Type 5 cyclic nucleotide phosphodiesterase inhibitors


Yonkenafil Hydrochloride
| Molecular Weight | 524.08 | |
| Formula | C24H33N5O4S • HCl |
804518-63-6 (Yonkenafil);
804519-64-0 (Yonkenafil Hydrochloride);
4H-Pyrrolo[2,3-d]pyrimidin-4-one, 2-[2-ethoxy-5-[(4-ethyl-1-piperazinyl)sulfonyl]phenyl]-3,7-dihydro-5-methyl-7-propyl-, hydrochloride (1:1)
2- [2-ethoxy –5- (4 – ethylpiperazine -1– sulfonyl) phenyl] -5 – methyl – 7 – n-Propyl-3 7 – PYRROLINE [2, 3 – d] pyrimidin – 4 – one
Yonkenafil hydrochloride, useful for treating erectile dysfunction and other PDE-5 mediated diseases eg female sexual dysfunction, benign prostatic hyperplasia, hypertension, allergic asthma, bronchitis, glaucoma, gastrointestinal motility disorders or Alzheimer’s Ydisease.
Yangtze River Pharmaceutical, under license from Jilin University, is developing yonkenafil (appears to be first disclosoed in WO2004108726), a PDE-5 inhibitor, for treating male erectile dysfunction.
In June 2016, yonkenafil was reported to be in phase 2 clinical development.
Yonkenafil hydrochloride is in phase II clinical trials for the treatment of erectile dysfunction (ED).
The compound was co-developed by Yangtze River Pharmaceutical and Tianjin Tasly Pharm.
Yonkenafil is a novel phosphodiesterase type 5 (PDE5) inhibitor. Here we evaluated the effect of yonkenafil on ischemic injury and its possible mechanism of action. Male Sprague-Dawley rats underwent middle cerebral artery occlusion, followed by intraperitoneal or intravenous treatment with yonkenafil starting 2h later. Behavioral tests were carried out on day 1 or day 7 after reperfusion. Nissl staining, Fluoro-Jade B staining and electron microscopy studies were carried out 24h post-stroke, together with an analysis of infarct volume and severity of edema. Levels of cGMP-dependent Nogo-66 receptor (Nogo-R) pathway components, hsp70, apaf-1, caspase-3, caspase-9, synaptophysin, PSD-95/neuronal nitric oxide synthases (nNOS), brain-derived neurotrophic factor (BDNF)/tropomyosin-related kinase B (TrkB) and nerve growth factor (NGF)/tropomyosin-related kinase A (TrkA) were also measured after 24h. Yonkenafil markedly inhibited infarction and edema, even when administration was delayed until 4h after stroke onset. This protection was associated with an improvement in neurological function and was sustained for 7d. Yonkenafil enlarged the range of penumbra, reduced ischemic cell apoptosis and the loss of neurons, and modulated the expression of proteins in the Nogo-R pathway. Moreover, yonkenafil protected the structure of synapses and increased the expression of synaptophysin, BDNF/TrkB and NGF/TrkA. In conclusion, yonkenafil protects neuronal networks from injury after stroke.

Erectile dysfunction (Erectile dysfunction, ED) refers to the duration can not be achieved, and (or) maintain an erection sufficient for satisfactory sexual life. ED can be divided according to different causes psychogenic, organic and mixed three categories, which are closely related to the aging process, but it is not inevitable disease with age.
The primary risk factors for ED include: high blood pressure, high cholesterol, diabetes, coronary and peripheral vascular disease, spinal cord injury or pelvic organs or surgery. According to statistics worldwide about 150 million men suffer from varying degrees of ED, 2025 the number of patients will double. More ED treatment options, such as oral medications phosphodiesterase 5 (PDE5) inhibitors, dopaminergic activator, a receptor blocker, intracavernous injection therapy, vacuum devices treatment, penile prosthesis treatment Wait. Wherein the selective phosphodiesterase 5 (PDE5) inhibitors are the most sophisticated study based on ED treatment, clinical treatment for ED is the first-line drugs. Has now approved the listing of these drugs were five sildenafil (Sildenafil), Tadalafil (Tadalafil), vardenafil (Vardenafil), to that of non-black (Udenafil) and Miro that non-( Mirodenafil).
In 2004 the Chinese patent CN03142399. X discloses a series pyrrolopyrimidine ketone compound of the structure and for the treatment of sexual dysfunction in animals, including humans, in particular male erectile dysfunction and TOE5 function-related diseases use; wherein the compound 1-HC1, i.e. 2- [2_ ethoxy-5- (4-ethyl-piperazine-1-sulfonyl) phenyl] -5-methyl-7-n-propyl -3 , 7-dihydro-pyrrolo [2, 3-d] pyrimidine-4-one monohydrochloride salt has been used as CN03142399. X Example features are disclosed compound named hydrochloride that non-gifted grams. This patent only to the preparation of the compounds have been described
PATENT
WO2004108726
http://www.google.co.in/patents/WO2004108726A1?cl=en
Example 1
Preparation of 2-[2-ethoxyl-5-(4-ethylpiperazinyl-1-sulfonyl)phenyl] -5-methyl-7-n-propyl-3,7-dihydropyrrolo[2,3-d]pyrimidin-4-one, its monohydrochloride and dihydrochloride
Route of synthesis
-


- (1a)2-amino-3-cyano-4-methylpyrrole;
- (1b)N-propyl-2-amino-3-cyano-4-methylpyrrole;
- (2)2-ethoxylbenzoyl chloride;
- (3a)N-(3-cyano-4-methyl-1H-pyrrol-2-yl)-2-ethoxylbenzamide;
- (3b)N-(3-cyano-4-methyl-1-n-propyl-1H-pyrrol-2-yl)-2-ethoxylbenzami de;
- (4a) 2-(2-ethoxylbenzamido)-4-methyl-1H-pyrrolo-3-formamide;
- (4b) 2-(2-ethoxylbenzamido)-4-methyl-1-n-propyl-1H-pyrrolo-3-formamide;
- (5) 2-(2-ethoxylphenyl)-5-methyl-3,7-dihydro-pyrrolo[2,3-d]pyrimidin -4-one;
- (6)2-(2-ethoxylphenyl)-5-methyl-7-n-propyl-3,7-dihydropyrrolo[2,3-d ]pyrimidin-4-one;
- (7)4-ethoxyl-3-(5-methyl-4-oxy-7-n-propyl-3,7-dihydropyrrolo[2,3-d] pyrimidin-2-yl)benzenesulfonyl chloride;
- (8)2-[2-ethoxyl-5-(4-ethylpiperazinyl-1-sulfonyl)phenyl]-5-methyl-7 -n-propyl-3,7-dihydropyrrolo[2,3-d]pyrimidin-4-one.


Preparation 1N-(3-cyano-4-methyl-1H-pyrrol-2-yl)-2-ethoxylbenzamide (3a) and N-(3-cyano-4-methyl-1-n-propyl-1H-pyrrol-2-yl)-2-ethoxylbenzamide (3b)
2-ethoxyl benzoic acid (10.0g, 60.2mmol) was added into thionyl chloride (20ml), and the mixture was refluxed with agitation for 40 minutes, and the excess amount of thionyl chloride was evaporated under reduced pressure. The residual was dissolved into dichloromethane (150ml). Within 30 minutes and being stirred on ice bath, the afore-obtained solution of 2-ethoxyl benzoyl chloride was dropped into the compound (1a) (7.0g, 56.8mmol) dissolved in tetrahydrofuran (80ml) and triethylamine (8.5ml, 61.0mmol). After completion, the mixture was stirred for 1 hour at 0°C . After being washed with water and filtrated with diatomaceous earth, the reaction solution was mixed with 20g of silica gel and evaporated to dryness. The resulting residual was eluted with dichloromethane by using silica gel(80g) column to obtain 7.5g of solid product (3a) with the yield of 48%. Furthermore, the sample for analysis was prepared by column chromatography (developing agent: dichloromethane: n-hexane=1:2) and recrystallization (dichloromethane: n-hexane=1:5).
mp 183~184°C (sublimation 162°C);\
IR (cm-1) : 3326, 3309, 2981, 2938, 2915, 2854, 2208, 1647, 1593, 1471, 1309, 1302, 1232, 1039, 923, 727, 655, 648;1H NMR (CDCl3) : δ 1.70 (t, J=7.0Hz, 3H), 2.15 (s, 3H), 4.32 (q, J=7.0Hz, 2H), 6.24 (s, 1H), 7.04 (d, 1H), 7.10 (m, 1H), 7.51 (dd, 1H), 8.20 (dd, J=7.9 and 1.8Hz, 1H), 10.69 (brs, 1H), 10.80 (s, 1H);13CNMR (CDCl3) : δ (CH3) 10.6, 15.0; (CH2) 65.7; (CH) 110.3, 112.3, 121.4132.1, 134.2; (C) 78.7, 115.6, 119.2, 119.4, 136.7, 157.0, 163.2;
MS (ES+) : m/z 287 (M+NH4) .
Elemental analysis (C15H15N3O2) : C 66.90%; H 5.61%; N 15.60%; 0 11.88%. The compound (3b) was prepared from compound (1b) according to the above-mentioned method with the yield of 41%.
mp 58~61°C;
IR (cm-1) : 3596, 3336, 2969, 2937, 2877, 2216, 1676, 1658, 1603, 1571, 1537, 1475, 1431, 1292, 1232, 1122, 1037, 927, 789, 752, 577;1H NMR (CDCl3): δ 0.88 (t, J=7.4Hz, 3H), 1.58 (t, J=7.0Hz, 3H), 1.75(m, 2H), 2.16 (s, 3H), 3.73 (t, J=7.4Hz, 2H),4.30 (q, J=7.0Hz, 2H), 6.36 (s, 1H), 7.04 (d, 1H), 7.11 (m, 1H), 7.48 (dd, 1H), 8.23 (dd, J=7.9 and 1.8Hz, 1H), 9.62 (brs, 1H) ;13C NMR (CDCl3) : δ (CH3) 11.1, 14.8; (CH2) 23.6, 48.3, 65.2; (CH) 112.5,117.0, 121.3, 132.5, 134.1; (C) 89.2, 115.6, 119.8, 120.5, 131.2, 157.1, 165.0;MS (ES+): m/z 329 (M+NH4).
Preparation 2
2-(2-ethoxylbenzamido)-4-methyl-1H-pyrrolo-3-formamide (4a) and 2-(2-ethoxylbenzamido)-4-methyl-1-n-propyl-1H-pyrrolo-3-formamide(4 b);
A mixture of N-(3-cyano-4-methyl-1H-pyrrol-2-yl)-2-ethoxylbenzamide(3a) (2.00g, 7.44mmol) or N-(3-cyano-4-methyl-1-n-propyl-1H-pyrrol-2-yl)-2 -ethoxylbenzamide(3b) (2.30g, 7.44mmol) of preparation 1 and 85% phosphoric acid (14.8ml) was stirred for 20 minutes at 130°C, cooled and poured into crushed ice (80g). The precipitations were filtrated and dried to give dark red solid of compound (3a) or (3b) with the yield of 80%. The product(3a) and (3b) of this step may be directly used for the next step without further purification.
Preparation 32-(2-ethxoylphenyl)-5-methyl-3,7-dihydropyrrolo[2,3-d]pyrimidin-4-one(5) and 2-(2-ethoxylphenyl)-5-methyl-7-n-propyl -3,7-dihydropyrrolo[2,3-d]pyrimidin-4-one(6)
A mixture of 2-(2-ethoxylbenzamido)-4-methyl-1H-pyrrolo-3-formamide (4a) (7.0g, 25.5mmol) of preparation 2 and dimethyl cyclohexylamine (20ml) was refluxed with agitation for 11 hours in N,N-dimethyl formamide (100ml). After evaporation the solvent by distillation under reduced pressure, the residual was extracted with dichloromethane, and the dichloromethane extraction was washed with water. the resultant extraction was dried with anhydrous sodium sulfate. n-hexane (80ml) was added into the residual and ground to give product (5) (6.0g) by filtration with the yield of 91%.
mp 219~221°C
IR (cm-1) : 3187, 3114, 3062, 2978, 2923, 1658, 1587, 1460, 1321, 1292, 1250, 1044, 771, 763;
1H NMR (DMSO-d6) : δ 1.35 (t, J=6.9Hz, 3H), 2.29 (s, 3H), 4.13 (q, J=7.0Hz, 2H), 6.79 (s, 1H), 7.05 (t, 1H), 7.14 (d, 1H), 7.45 (dd, 1H), 7.76 (dd, 1H), 11.35 (brs, 1H), 11.54 (brs, 1H);
13C NMR (DMSO-d6) : δ (CH3) 11.2, 14.5; (CH2) 64.2; (CH) 113.0, 118.0, 120.6, 130.1, 131.9, (C) 105.0, 113.6, 121.9, 148.5, 149.8, 156.5, 159.2; MS(ES+) : m/z 287 (M+NH4) .
The compound (6) was prepared from compound(4b) according to the above-mentioned method with the yield of 85%
mp 124~127°C
IR (cm-1) : 3234, 3184, 3141, 3103, 3056, 2956, 2943, 2869, 1654, 1595, 1567, 1468, 1311, 1267, 1243, 1191, 1118, 1047, 758;
1H NMR (CDCl3) : δ 0.88 (t, J=7.5Hz, 3H), 1.23 (t, 3H), 1 . 80 (q, 2H), 2. 42 (s, 3H), 4.08 (t, J=7.2Hz, 2H), 4.22 (q, 2H), 6.60 (s, 1H), 7.01 (d, J=8.3Hz, 1H), 7.08 (t, 1H), 7.40 (m, 1H), 8.35 (dd, J=8.0 and 1.9 Hz, 1H), 11.02 (brs, 1H).
Preparation 42-(2-ethxoylphenyl)-5-methyl-7-n-propyl-3,7-dihydro-pyrrolo[2,3-d] pyrimidin-4-one(6):
A mixture of compound (5) (1.5g, 5.57mmol) of preparation 3, n-propyl bromide (2.0g, 16.3mmol) and potassium carbonate (5g, 36.2mmol) was dissolved in acetone (15ml), refluxed with agitation by heating for 15 hours, after the solids were filtrated out, the filtrate was dried under reduced pressure. The resultant was developed by column chromatography, using dichloromethane as mobile phase to obtain 0.6g of product (6) with yield of 35%. The physical/chemical data were identical with that of the above-mentioned.
Preparation 54-ethoxyl-3-(5-methyl-4-oxy-7-n-propyl-4,7-dihydropyrrolo[2,3-d] pyrimidin-2-yl)benzenesulfonyl chloride(7):
2-(2-ethxoylphenyl)-5-methyl-7-n-propyl-3,7-dihydropyrrolo[2,3-d] pyrimidin-4-one(6) (1.25g, 4.01mmol) of preparation 4 was added into chlorosulfonic acid (4ml) that was dissolved in acetic ether (20ml), stirred at 0°C by two batches. The obtained solution was stirred at 0 °C for 30 minutes, and then reacted with agitation at room temperature for 3 hours. The resultant solution was poured into the a mixture of icy water (50ml) and acetic ether (50ml) . The organic layer was separated, washed with cold water (5ml), desiccated with anhydrous sodium sulfate and concentrated to dryness to afford 1.33g of product as yellow foam. The yield was 81%. The product was used directly for the next reaction.
Compound 1:
BASE
2-[2-ethoxyl-5-(4-ethyl-piperazinyl-1-sulfonyl)phenyl]-5-methyl-7-n -propyl-3,7-dihydropyrrolo[2,3-d]pyrimidin-4-one (8):
4-ethoxyl-3-(5-methyl-4-oxy-7-n-propyl-4,7-dihydro-3H-pyrrolo[2,3-d ]pyrimidin-2-yl)benzenesulfonyl chloride(7) (1.00g, 2.44mmol) of Preparation 5 was dissolved into dichloromethane (20ml), stirred at 0 °C, into which 1-ethyl piperazine (0.78ml, 6.10mmol) was added slowly. Reactant solution was stirred at 0°C for 5 minutes, and then sequentially stirred at room temperature for 5 hours. The crude product was washed with water and dried with anhydrous sodium sulfate to give 1. 2g of product as yellow foam. Continuously, the product was refined by column chromatography (acetic ether: methanol=20:1) to afford 0.89g of product as a yellow solid with yield of 75%.
mp: 174~176°C (EtOAc);
IR (cm-1) : 3324, 2960, 2923, 2869, 2862, 2767, 1682, 1560, 1458, 1355, 1282, 1247, 1172, 1149, 739, 615, 588, 555;
1H NMR(CDCl3) : δ 0.89(t,J=7.4Hz, 3H), 0.99(t, J=7.2Hz, 3H), 1.61(t,J=7.0Hz,3H),1.77-1.86(m, 2H), 2.35(m, 2H), 2.41(s, 3H), 2.50(brs, 4H), 3.05(brs,4H), 4.08(t, J=7.0Hz, 2H), 4.29-4.37(q, 2H), 6.61(s, 1H), 7.11(d, J=8.8Hz,1H), 7.77(dd, J=8.7 and2.2Hz, 1H), 8.74(d, J=2.2, 1H), 10.63(brs, 1H);
13C NMR(CDCl3) : δ (CH3) 11.0, 11.3, 11.8, 14.3; (CH2)23.8, 45.9, 46.1, 51.6, 51.7, 65.8; (CH)112.9, 121.1, 130.6, 131.3;(C)105.7,114.6, 121.4, 127.8, 146.8, 147.3, 159.3, 159.6;MS(ES+): m/z 505(M+NH4).
Elemental analysis (C24H33N5O4S) : theoretical value C 59.12%; H 6.82%; N 14.36%; practically measured value C59.38%; H 7.10%; N 14.12%.
Compound 1-HCl:
2-[2-ethoxyl-5-(4-ethylpiperazinyl-1-sulfonyl)phenyl]-5-methyl-7-n-propyl-3,7-dihydropyrrolo[2,3-d]pyrimidin-4-one monohydrochloride (9) :
The free alkali (compound 1) (1.00g, 2.05mmol) was dissolved into ether (10ml) and dichloromethane (10ml), into which the solution of 4M hydrochloric acid (HC1)- dioxane (0.51ml, 2.04mmol) diluted with ethyl ether (10ml) was dropped with agitation. After completion, the resulting solution was continued to stir at room temperature for 20 minutes, filtrated and dried to give 1.01g of monohydrochloride with yield of 94%.
mp: 147~150°C;
IR(cm-1): 2964, 2931, 2675, 2599, 2462, 1668, 1574, 1456, 1348, 1167, 933, 588;
1H NMR(D2O): δ 0.72(m, 3H),1.24(t, J=7.3Hz, 3H), 1.45(m, 3H), 1.59(m, 2H), 2.04(s, 3H), 2.77-3.81(all brs, 8H), 3.20(q, 2H), 3.75(m, 2H), 4.20(m, 2H), 6.62(m, 1H), 7.17(m, 1H), 7.73(m, 1H), 8.22(s, 1H).
Elemental analysis (C24H33N5O4S. HCl) : theoretical value C 55.00%; H 6.54%; N 13.36%; practically measured value C55.28%; H 6.41%; N 13.07%.
PATENT
WO 2016095650
https://patentscope.wipo.int/search/en/detail.jsf?docId=WO2016095650&redirectedID=true
SEE
https://www.google.com/patents/CN1552714A?cl=en
Spectral Analysis
13C NMR PREDICT
COSY PREDICT
| CN1552714A * | Jun 6, 2003 | Dec 8, 2004 | 天津倍方科技发展有限公司 | 2-substituted benzyl-5,7-dihydrocarbyl-3,7-dihydro pyrroline [2,3-d] pyromidine-4-one derivative ,its preparation and medicinal use |
| CN102970965A * | Apr 4, 2011 | Mar 13, 2013 | Sk化学公司 | Composition containing PDE5 inhibitor for relieving skin wrinkles |
| WO2007067570A1 * | Dec 5, 2006 | Jun 14, 2007 | Biomarin Pharmaceutical Inc. | Methods and compositions for the treatment of disease |
//////////yonkenafil, Phase 2, Erectile dysfunction , phosphodiesterase type 5 (PDE5) inhibitor, Tasly Pharmaceutical Group; Yangtze River Pharmaceutical Group
Cc4cn(CCC)c1c4N/C(=N\C1=O)c2cc(ccc2OCC)S(=O)(=O)N3CCN(CC3)CC
Gisadenafil


