Home » Posts tagged 'world drug tracker' (Page 5)
Tag Archives: world drug tracker
 DRUG APPROVALS BY DR ANTHONY MELVIN CRASTO
.....FOR BLOG HOME CLICK HERE
Flag and hits
ORGANIC SPECTROSCOPY

SUBSCRIBE

 Subscribe in a reader
Enter your email address:
Delivered by FeedBurner
Subscribe to New Drug Approvals by Email


Argatroban

Argatroban
Molecular Formula: C23H36N6O5S
Formula Weight: 508.63
CAS No.: 74863-84-6
(2R,4R)-1-[(2S)-5-(diaminomethylideneamino)-2-
[[(3R)-3-methyl-1,2,3,4-tetrahydroquinolin-8-yl]
sulfonylamino]pentanoyl]-4-methyl-piperidine-2-
carboxylic acid
PATENT
US 7,589,106, 7,687,516, EP 0008746; US 4258192, US 4201863
Argatroban is an anticoagulant that is a small molecule direct thrombin inhibitor.[1] In 2000, argatroban was licensed by the Food and Drug Administration (FDA) for prophylaxis or treatment of thrombosis in patients with heparin-induced thrombocytopenia (HIT). In 2002, it was approved for use during percutaneous coronary interventions in patients who have HIT or are at risk for developing it. In 2012, it was approved by the MHRA in the UK for anticoagulation in patients with Heparin-Induced Thrombocytopenia Type II (HIT) who require parenteral antithrombotic therapy.[2]
Argatroban is given intravenously and drug plasma concentrations reach steady state in 1-3 hours.[3] Argatroban is metabolized in the liver and has a half-life of about 50 minutes. It is monitored by PTT. Because of its hepatic metabolism, it may be used in patients with renal dysfunction. (This is in contrast to lepirudin, a direct thrombin inhibitor that is primarily renally cleared).

Argatroban is used as an anticoagulant in individuals with thrombosis and heparin induced thrombocytopenia. Often these individuals require long term anticoagulation. If warfarin is chosen as the long term anticoagulant, this poses particular challenges due to the falsely elevated prothrombin time and INR caused by argatroban. The combination of argatroban and warfarin may raise the INR to greater than 5.0 without a significant increased risk of bleeding complications.[4] One solution to this problem is to measure the chromogenic factor X level. A level < 40-45% typically indicates that the INR will be therapeutic (2-3) when the argatroban is discontinued.
……………………………………………………….
Argatroban monohydrate

Argatroban is a synthetic direct thrombin inhibitor and the chemical name is 1-[5-[(aminoiminomethyl) amino]-1-oxo2[[(1,2,3,4-tetrahydro-3-methyl-8-quinolinyl)sulfonyl]amino]pentyl]-4-methyl-2- piperidinecarboxylic acid, monohydrate. Argatroban has 4 asymmetric carbons. One of the asymmetric carbons has an R configuration (stereoisomer Type I) and an S configuration (stereoisomer Type II). Argatroban consists of a mixture of R and S stereoisomers at a ratio of approximately 65:35.
The molecular formula of argatroban is C23H36N6O5S•H2O. Its molecular weight is 526.66 g/mol. cas 141396-28-3
Argipidine, Argatroban monohydrate, GN1600, DK-7419, MDI-805 Acova, Slonnon, Novastan
Mitsubishi Chemical (Originator), Encysive Pharmaceuticals (Licensee), Mitsubishi Pharma (Distributor), Daiichi Pharmaceutical (Codevelopment), GlaxoSmithKline (Codevelopment), Mitsubishi Pharma (Codevelopment), Sanofi-SynthLabo (Codevelopment)
Antithrombocytopenic, CARDIOVASCULAR DRUGS, Cerebrovascular Diseases, Treatment of, HEMATOLOGIC DRUGS, Hematopoiesis Disorders Therapy, Ischemic Stroke, Treatment of, NEUROLOGIC DRUGS, Peripheral Vascular Disease, Treatment of, Stroke, Treatment of, Treatment of Peripheral Obstructive Vascular Disease, Thrombin Inhibitors
Synthesis of argatroban on the method reported in the literature there are two synthetic routes, patent EP8746, US4258192, US4201863, JP8115267 relates to a route is: with 4 – methyl-piperidine as a starting material was prepared first intermediate body (2R, 4R) -4 – methyl-2 – ethyl-piperidine, and the first and a t-BOC protected amino nitro-L-arginine condensation, and then the 3 – methyl – 8 – quinoline sulfonyl chloride condensation after hydrolysis, hydrogenation, hydration be argatroban. This entry route synthesis process complicated procedure to be carried out under the protection of nitrogen, the raw material is highly toxic gas phosgene, the operation more difficult.
US4117127, JP02-212473, EP823430, EP8746, JC S Perk Transl 1981 (5), JP02-212473 relates to an alternative route is: nitro L-arginine prior to the 3 – methyl-8 – subsequent condensation quinoline sulfonyl chloride Intermediate (2R, 4R) -4 – methyl-2 – piperidinecarboxylate condensation, and then after hydrolysis, hydrogenation, hydration be argatroban. This synthetic route despite the relatively simple process method, to obtain raw materials, but this method using reagents such as phosphorus oxychloride, phosphorus trichloride has a pungent odor, easy to absorb moisture in humid air, intense smoke, environmental pollution, greater stimulation of the body’s respiratory tract, can cause eye and skin irritation and burning, and the use of this method, complex operation, low yield, high cost.
Patent CN100586946C Argatroban discloses a method for separating optically active isomeric compounds, the feedstock argatroban mixed solvent of alcohol and water was heated to reflux 5-10 hours, cooled and allowed to stand, and filtered to give White crystalline product, dried, repeated 2-6 times. [0008] Patent CN101033223A discloses a Argatroban is the main by-product (2R, 4R)-l_ [N2-(3_ methyl-8 – quinolinesulfonyl)-L-arginyl] -4 – methyl-2 – carboxylic acid, argatroban, and the byproducts are difficult to isolate, argatroban two diastereomeric isomer 21 (S) and 21 (R) separation of work attracted a lot of research persons. Because both physical and chemical properties are very similar, so separation is very difficult. 1993 Rawson, Thomas E.; VanGorp, Kimmie A.; Yang, Janet so first by high pressure liquid chromatography and column chromatography separation to obtain a single 21 (S) and 21 (R) argatroban [ Journal of Pharmaceutical Sciences vol. 82, No. 6,672]; Thibaudeau Karen et al. reported Protein A chromatographic separation [US6440417]. However, due to the separation of these methods a small amount of low efficiency, so there is no practical value industrialization. 2006 China Tianjin Weijie Technology Co., Ltd. Song Honghai et al. Reported using recrystallization Separation 21 (S) and 21 (R) argatroban way [0 With 951,936 it], so that the mass 21 (5) Aga music classes as possible, but the law of low yield, complicated operation, high cost, and a large amount of a small amount of 21 (S) of 21 (R)-product argatroban, from the viewpoint of industrial production, is still a ideal method. [0009] These methods can be effectively prepared argatroban, but the purity of the desired product is not high, poor color, content is low, affecting the quality of the results of its preparation.
U.S. Pat. No. 4,201,863 (6 May 1980) and EP 8746 (filed on 22 Aug. 1979 with priority based on the application for the cited US patent) describe a class of N2-arylsulphonyl-L-argininamide drugs, with anti-thrombotic activity, and the processes for obtaining them. Of these, the compound 4-methyl-1-[N2-(3-methyl-1,2,3,4-tetrahydro-8-quinolinesulphonyl)-L-arginyl]-2-piperidine carboxylic acid (argatroban, isomers mixture) is described. The described process comprises the synthesis of an intermediate NG-substituted-N2-quinolinesulphonyl-L-argininamide from which the desired compound is obtained by catalyzed hydrogenolysis or acidolysis and catalyzed hydrogenation. The general conditions provided for the hydrogenolysis and hydrogenation reaction are: i) inert solvents (methanol, ethanol, tetrahydrofuran or dioxane); ii) presence of a catalyst (Raney nickel, palladium, platinum, ruthenium, rhodium); iii) hydrogen atmosphere at a pressure between 1 and 100 kg/cm2 and preferably between 5 and 50 kg/cm2; iv) temperature between 0° C. and 200° C. and preferably between 50° C. and 150° C.; v) reaction temperature from 2 hours to 120 hours. The crude product obtained is then purified by trituration or by re-crystallization from diethyl ether-tetrahydrofuran, diethyl ether-methanol or from water-methanol or by chromatography. No example is given of this purification step. In particular, both U.S. Pat. No. 4,210,863 and EP 8746 in example 1(E) describe the preparation of argatroban, isomers mixture. This compound is obtained in amorphous form by hydrogenation of [NG-nitro-N2-(3-methyl-8-quinolinesulphonyl)-L-arginyl]-4-methyl-2-piperidine carboxylic acid in ethanol in the presence of Pd/C with hydrogen pressure of 10 kg/cm2 at 100° C. for 8 hours. The catalyst is removed by filtration of the ethanol solution which is then evaporated without further purification and/or re-crystallization steps. In the US patent at issue as indeed in patent application EP 8746, no mention is made of polymorphic forms of the compounds and, for the obtained compound, the following characteristics are reported: Amorphous solid, I.R. (KBr) (cm−1) 3400; 1620; 1460; 1380; Molecular composition (%): theoretical C 54.31; H 7.13; N 16.52; found (%) C 54.01; H 6.98; N 16.61.
U.S. Pat. No. 4,258,192 (24 Mar. 1981) (continuation-in-part of the aforesaid patent application U.S. Pat. No. 4,201,863) and the same patent application EP 8746 describe the stereoisomers and the preparation thereof, including argatroban used as an active principle in medicaments, i.e. the stereoisomer (2R,4R)-4methyl-[4N2-(3-methyl-1,2,3,4-tetrahydro-8-quinolinesulphonyl)-L-arginyl]-2-piperidine carboxylic acid, with the following characteristics: melting point (m.p.). 188-191° C.; I.R. (KBr) (cm−1) 3400, 1620, 1460, 1380; Molecular composition (%): theoretical C 54.31; H 7.13; N 16.52; found (%) C 54.05; H 6.94; N 16.65. The compound is prepared according to the description given in examples 1(E) in U.S. Pat. No. 4,258,192 and 2(E) and 3 in EP 8746 respectively by hydrogenation of (2R,4R) 1-[NG-nitro-N2-(3-methyl-1,2,3,4-tetrahydro-8-quinolinesulphonyl)-L-arginyl]-2-piperidine carboxylic acid in ethanol in presence of acetic acid catalyzed by Pd/C. After filtering the mass to remove the catalyst, the solvent is evaporated and the residue suspended in chloroform, the solution treated with a saturated sodium bicarbonate solution or 1N sodium hydroxide solution and after washing, the solvent is evaporated. The compound is then re-crystallized from ethanol. Again in this case, no reference is made to the obtainment of monohydrate polymorphic forms.
Said polymorphic forms are described instead in the publication Biochem. Biophys. Res. Comm. 1981, 101, 440-446 in the context of stereoisomer preparation. The monohydrate polymorph of the (2R,4R) stereoisomer is prepared by re-crystallization from ethanol/water and the reported characteristics are: m.p. 176-180° C.; [α]D 27 +76.1° (c 1, 0.2N HCl).
U.S. Pat. No. 5,925,760 (20 Jul. 1999) and EP 0823430 (filed 4 Aug. 1997) subsequently describe a new method for preparing argatroban by means of a new intermediate N2-(3-methyl-8-quinolinesulphonyl)-NG-nitro-L-arginine. In particular the patent makes reference to the preparation of a crystalline monohydrate form of argatroban, referring back to examples (D) and (E) of Japanese patent publication No. (Hei)-2-31055/1990 and generically to an I.R. spectrum identical to that of the commercially available argatroban compound. The relevant example in the cited patent publication is example (E), while example (D) concerns the preparation of (2R,4R)-1-[NG-nitro-N2-(3-methyl-8-quinolinesulphonyl)-L-arginyl]-4-methyl-2-piperidine carboxylic acid. This compound represents the starting compound for argatroban preparation by catalytic reduction in the presence of Pd/C. The crude argatroban obtained is then purified by extraction with chloroform, treatment with a saturated sodium bicarbonate solution and, after solvent evaporation, re-crystallization from ethanol or from 15% alcohol in water. It should be noted however that the Japanese patent makes no mention of the monohydrate form of argatroban being obtained and that for the compound the following characteristics are reported: m.p. 188-191° C.; molecular composition (theoretical/found) (%): C 54.31/54.01; H 7.13/6.98; N 16.52/16.61; I.R. (KBr) (cm−1) 3400; 1620; 1460; 1380. These analytical data, with the exception of the unreported melting point, are the same as those indicated in the cited patent documents describing a mixture of (2R,4R)-4methyl-[4N2-(3S-methyl-1,2,3,4-tetrahydro-8-quinolinesulphonyl)-L-arginyl]-2-piperidine carboxylic acid and (2R,4R)-4methyl-1-[N2-(3R-methyl-1,2,3,4-tetrahydro-8-quinolinesulphonyl)-L-arginyl]-2-piperidine carboxylic acid isomers of argatroban, but do not correspond to the melting point given in the publication, being the only document that identifies the monohydrate form of argatroban.
More recently, patent application CN 1,951,937 (filing date 10 Nov. 2006) described a method for preparing hydrated argatroban by treating argatroban with large quantities of water (more than 60 and up to 80 volumes of distilled water per gram of argatroban) at a temperature of 80-100° C. for a time of 0.5-1 hour and crystallization by cooling. The water content reported is comprised between 3.3 and 3.8% and the ratio of dextroisomer R to levoisomer S is R:S=63-67: 37-33.
Argatroban is a compound of wide therapeutic use, for which reason the need still exists to provide a compound of pharmaceutically acceptable quality obtained by easily industrialized and economically convenient methods. With regard to the monohydrate, this form is preferable for the applicative purpose since the anhydrous form is unstable and tends to become hydrated and/or wet. Moreover it crystallizes only with difficulty at the correct ratio between the diastereoisomers.

…..

the protection of 4-methylpiperidine (I) with (Boc)2O gives the carbamate (II), which is condensed with benzyl chloroformate by means of sec-butyl lithium and TMEDA in ethyl ether to yield (?-trans-1-(tert-butoxycarbonyl)-4-methylpiperidine-2-carboxylic acid benzyl ester (III). Deprotection of the NH group of (III) with HCl in ethyl acetate affords (?-trans-4-methylpiperidine-2-carboxylic acid benzyl ester (IV), which is condensed with the protected arginine derivative (V) by means of isobutyl chloroformate and TEA to provide the corresponding amide as a diastereomeric mixture. Resolution of this mixture by flash chromatography furnishes the desired diastereomer (VI), which is treated with HCl in ethyl acetate in order to remove the Boc-protecting group to yield compound (VII). Condensation of compound (VII) with 3-methylquinoline-8-sulfonyl chloride (VIII) by means of TEA in dichloromethane affords the expected sulfonamide (IX). Finally, this compound is submitted to hydrogenation with H2 over Pd/C in AcOH/ethanol in order to produce debenzylation, cleavage of the NO2 group and hydrogenation of the pyridine ring to yield argatroban.
………….
Argatroban, i.e., (2R,4R)-1-((2S)-5-((Aminoiminomethyl)amino)-1-oxo-2-((1,2,3,4-tetrahydro-3-methyl-8-quinolinyl)sulfonyl)amino) pentyl)-4-methyl-2-piperidine carboxylic acid, has two diastereoisomers: 21(R) and 21(S). Usually the ratio of 21(R) to 21(S) is 64-65: 36-35 (U.S. Pat. No. 6,440,417, Cossy. J., et al, Bioorganic & Medicine Chemistry Letters, 11 (2001), 1989-1992, Journal of pharmaceutical Sciences, Vol. 82, No. 6, 672 (1993)).
The structure formula of Argatroban is reported below:

-
- 21(S) Argatroban, X=CH3, Y═H;
- 21(R) Argatroban, X=H, Y=CH3;
- Argatroban, 21(S): 21 (R)=35:65.
The chemical names of the two diastereoisomers mentioned above are:
- 21(S) Argatroban: (2R,4R)-1-((2S)-5-((Aminoiminomethyl)amino)-1-oxo-2-((((3S)-1,2,3,4-tetrahydro-3-methyl-8-quinolinyl)sulfonyl)amino)pentyl)-4-methyl-2-piperidine carboxylic acid (121785-72-6); and
- 21(R) Argatroban: (2R,4R)-1-((2S)-5-((Aminoiminomethyl)amino)-1-oxo-2-((((3R)-1,2,3,4-tetrahydro-3-methyl-8-quinolinyl)sulfonyl)amino)pentyl)-4-methyl-2-piperidine carboxylic acid (121785-71-5).
In 1978, S. Akamoto et al from Japanese Mitsubishi Chemical Corporation first disclosed the anti-thrombin activity of Argatroban monohydrate (U.S. Pat. No. 4,101,653). In the next 20 years, numerous researchers had in-depth studies on Argatroban about its biological activity and medicine values. In 1981, S. Akamoto compared Argatroban with heparin in vivo (Okamoto, S. et al., Biochem. Biophys. Res. Commun. 101, 440 (1981)); T. Kumoto disclosed its three-dimensional selective activity (Kumada, T. et al., Thromb. Res. 24, 285 (1981)). In 1984, R. Kumato made a clinical evaluation of hemodialysis of Argatroban (Kikumoto, R. et al., Biochemistry 23, 85 (1984)), and in 1986, he further disclosed that Argatroban can inhibit the thrombin activity of mammals, and can be used as active ingredient to treat and prevent thrombosis and as an inhibitor of platelet aggregation. Argatroban monohydrate can be used as a selective anti-thrombosis agent for treatment of chronic arterial blockage and cerebral thrombosis, etc (JP 61-48829). In 1992 and 1993, Taparelli and Jakubowski separately disclosed the reversibility of Argatroban in anti-thrombin (Taparelli, C., Trends Pharmacol. Sci., 1993, 14, 366, Jakubowski, J. A. et al, Rep. Med. Chem., 1992, 27, 99). In 1990s, many researchers such as L. R. Buch reported other related research (Buch, L. R., Cadiosvasc. Drug Rev., 1991, 9, 247, Strupcnewski, J. D. et al., Academic: San Diego, 1991; Vol. 26, p 299, Brundish, D. et al., J. Med. Chem. 1999; 42, 4584, Shebuski, R. J., Academic: San Diego, 1999; Vol. 26, p 98). In 1992, Argatroban monohydrate was first approved as an anti-thrombin medicine in Japan (Hijikata-Okunomiya, A., et al, Thromb. Hemostasis, 1992, 18, 135).
Updated in sept 2015, reader there may be duplication
| Argatroban |
| AC1L99H9; AC-15185; TL8005144; C04931; | |
| Molecular Formula: | C23H36N6O5S |
|---|---|
| Molecular Weight: | 508.63414 g/mol |
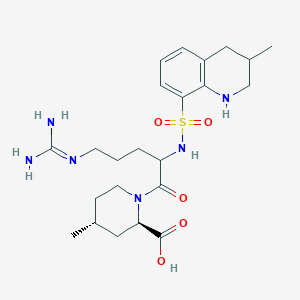
(2R,4R)-1-[5-(diaminomethylideneamino)-2-[(3-methyl-1,2,3,4-tetrahydroquinolin-8-yl)sulfonylamino]pentanoyl]-4-methylpiperidine-2-carboxylic acid, cas 74863-84-6
| Patent | Submitted | Granted |
|---|---|---|
| Prodrugs of (2R)-2-Propyloctanoic Acid For the Treatment of Stroke [US7495029] | 2008-06-05 | 2009-02-24 |
Tomiya Mano, Jin Shiomura, “Argatroban preparations for ophthalmic use.” U.S. Patent US5506241, issued October, 1986.
Argatroban is an anticoagulant that is a small molecule direct thrombin inhibitor.[1] In 2000, argatroban was licensed by the Food and Drug Administration (FDA) for prophylaxis or treatment of thrombosis in patients with heparin-induced thrombocytopenia (HIT). In 2002, it was approved for use during percutaneous coronary interventions in patients who have HIT or are at risk for developing it. In 2012, it was approved by the MHRA in the UK for anticoagulation in patients with Heparin-Induced Thrombocytopenia Type II (HIT) who require parenteral antithrombotic therapy.[2]
Argatroban is given intravenously and drug plasma concentrations reach steady state in 1-3 hours.[3] Argatroban is metabolized in the liver and has a half-life of about 50 minutes. It is monitored by PTT. Because of its hepatic metabolism, it may be used in patients with renal dysfunction. (This is in contrast to lepirudin, a direct thrombin inhibitor that is primarily renally cleared).
Argatroban is a direct, selective thrombin inhibitor. The American College of Cardiologists (ACC) recommend using bivalirudin or argatroban in patients who have had, or at risk for, heparin induced thrombocytopenia (HIT) and are undergoing percutaneous coronary intervention. Argatroban is a non-heparin anticoagulant shown to both normalize platelet count in patients with HIT and prevent the formation of thrombi. Parental anticoagulants must be stopped and a baseline activated partial thromboplastin time must be obtained prior to administering argatroban.
Transitioning to warfarin in individuals with heparin induced thrombocytopenia
Argatroban is used as an anticoagulant in individuals with thrombosis and heparin induced thrombocytopenia. Often these individuals require long term anticoagulation. If warfarin is chosen as the long term anticoagulant, this poses particular challenges due to the falsely elevated prothrombin time and INR caused by argatroban. The combination of argatroban and warfarin may raise the INR to greater than 5.0 without a significant increased risk of bleeding complications.[4] One solution to this problem is to measure the chromogenic factor X level. A level < 40-45% typically indicates that the INR will be therapeutic (2-3) when the argatroban is discontinued.

http://www.google.com/patents/WO2009124906A2?cl=en
………….
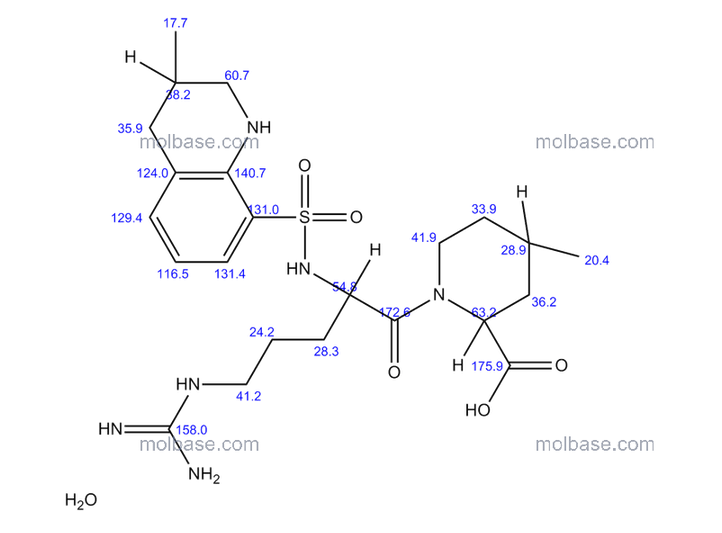
NMR paper
Complete 1H and 13C assignments of (21R) and (21S) diastereomers of argatroban
Article first published online: 20 DEC 2007
DOI: 10.1002/mrc.2122, http://onlinelibrary.wiley.com/doi/10.1002/mrc.2122/abstract


click on image for clear view
1H NMR PREDICT


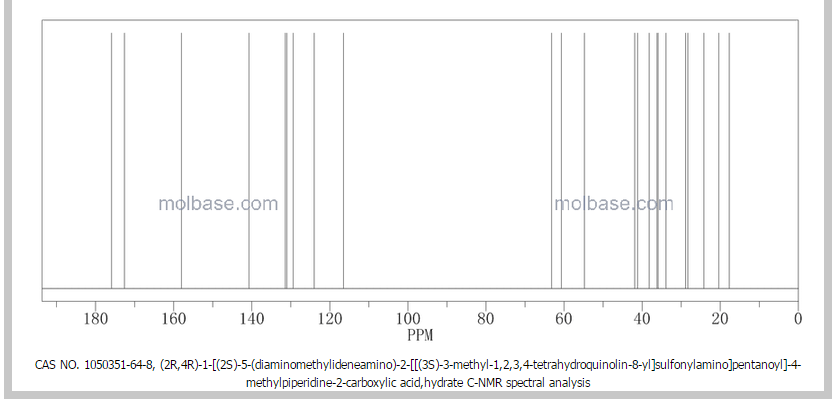
13C NMR PREDICT
References
1 Di Nisio M, Middeldorp S, Buller HR. Direct thrombin inhibitors. N Engl J Med 2005;353:1028-40. PMID 16148288
2http://www.pharmatimes.com/Article/12-07-03/UK_launch_for_Mitsubishi_s_blood_thinner_Exembol.aspx
3Dhillon S. Argatroban: A Review of its Use in the Management of Heparin-Induced Thrombocytopenia. Am J Cardiovasc Drugs 2009; 9 (4): 261-82. Link text
- Hursting MJ, Lewis BE, Macfarlane DE. (2005). “Transitioning from argatroban to warfarin therapy in patients with heparin-induced thrombocytopenia.”. Clin Appl Thromb Hemost 11 (3): 279–87. doi:10.1177/107602960501100306. PMID 16015413.
External links
| EP0823430A1 * | Aug 4, 1997 | Feb 11, 1998 | Mitsubishi Chemical Corporation | Method for preparing n2-arylsulfonyl-l-argininamides |
| US4201863 * | Aug 31, 1978 | May 6, 1980 | Mitsubishi Chemical Industries, Limited | N2 -Arylsulfonyl-L-argininamides and the pharmaceutically acceptable salts thereof |
| Reference | ||||
|---|---|---|---|---|
| 1 | None | |||
| 2 | * | OKAMOTO S ET AL: “Potent inhibition of thrombin by the newly synthesized arginine derivative No. 805. The importance of stereo-structure of its hydrophobic carboxamide portion” BIOCHEMICAL AND BIOPHYSICAL RESEARCH COMMUNICATIONS, ACADEMIC PRESS INC. ORLANDO, FL, US, vol. 101, no. 2, 30 July 1981 (1981-07-30), pages 440-446, XP024844713 ISSN: 0006-291X [retrieved on 1981-07-30] cited in the application | ||
| 3 | * | SONG H: “Method for preparing argatroban monohydrate in pure water” CASREACT,, 25 April 2007 (2007-04-25), XP002493272 & CN 1 951 937 A (TIANJIN WEIJIE TECHNOLOGY CO L [CN]) 25 April 2007 (2007-04-25) | ||
| Citing Patent | Filing date | Publication date | Applicant | Title |
| WO2012136504A1 | Mar 26, 2012 | Oct 11, 2012 | Lundbeck Pharmaceuticals Italy S.P.A. | Method for the preparation of process intermediates for the synthesis of argatroban monohydrate |
| CN102408468A * | Sep 20, 2011 | Apr 11, 2012 | 海南灵康制药有限公司 | Argatroban compound and preparation method thereof |
| EP2752412A1 | Mar 26, 2012 | Jul 9, 2014 | Lundbeck Pharmaceuticals Italy S.p.A. | Intermediates for the synthesis of Argatroban monohydrate |
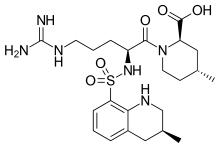
Argatroban is a synthetic direct thrombin inhibitor and the chemical name is 1-[5[(aminoiminomethyl)amino]1-oxo-2-[[(1,2,3,4-tetrahydro-3-methyl-8-quinolinyl)sulfonyl]amino]pentyl]-4methyl-2-piperidinecarboxylic acid, monohydrate. Argatroban has 4 asymmetric carbons. One of the asymmetric carbons has an R configuration (stereoisomer Type I) and an S configuration (stereoisomer Type II). Argatroban consists of a mixture of R and S stereoisomers at a ratio of approximately 65:35.
The molecular formula of argatroban is C23H36N6O5S•H2O. Its molecular weight is 526.66 g/mol. The structural formula is:
|
Argatroban Injection is a sterile, non-pyrogenic, clear, colorless to pale yellow isotonic solution. It is supplied in a single use polyolefin bag containing 250 mg of argatroban in 250 mL sodium chloride solution (1 mg/mL). Each mL contains 1 mg argatroban, 9 mg sodium chloride, USP, and 3 mg sorbitol, NF in water for injection, USP. The pH of the solution is between 3.2 to 7.5.
 |
|
| Systematic (IUPAC) name | |
|---|---|
|
(2R,4R)-1-[(2S)-5-(diaminomethylideneamino)-2-
[[(3R)-3-methyl-1,2,3,4-tetrahydroquinolin-8-yl] sulfonylamino]pentanoyl]-4-methyl-piperidine-2- carboxylic acid |
|
| Clinical data | |
| Trade names | Argatroban |
| AHFS/Drugs.com | monograph |
| Routes of administration |
intravenous |
| Pharmacokinetic data | |
| Bioavailability | 100% (intravenous) |
| Protein binding | 54% |
| Metabolism | hepatic |
| Biological half-life | 39 and 51 minutes |
| Identifiers | |
| CAS Registry Number | 74863-84-6  |
| ATC code | B01AE03 |
| PubChem | CID: 440542 |
| DrugBank | DB00278 |
| ChemSpider | 389444 |
| UNII | OCY3U280Y3 |
| KEGG | C04931 |
| ChEMBL | CHEMBL1166  |
| Chemical data | |
| Formula | C23H36N6O5S |
| Molecular mass | 508.635 g/mol |
//////
Drug spotlight- Zafirlukast

ZAFIRLIKAST 
cyclopentyl 3-{2-methoxy-4-[(o-tolylsulfonyl)carbamoyl]benzyl}-1-methyl-1H-indol-5-ylcarbamate 107753-78-6
Matassa, V.G. et al, J. Med. Chem., v. 33, 1781 (1990);
U. S. Patent No. 4,859,692;
U. S. Patent No. 5,993,859;
http://www.accessdata.fda.gov/drugsatfda_docs/label/2011/020547s031lbl.pdf
Zafirlukast is an oral leukotriene receptor antagonist (LTRA) for the maintenance treatment of asthma, often used in conjunction with an inhaled steroid and/or long-acting bronchodilator. It is available as a tablet and is usually dosed twice daily. Another leukotriene receptor antagonist is montelukast (Singulair), taken once daily. Zileuton (Zyflo), also used in the treatment of asthma via its inhibition of 5-lipoxygenase, is taken four times per day.
Zafirlukast blocks the action of the cysteinyl leukotrienes on the CysLT1 receptors, thus reducing constriction of the airways, build-up of mucus in the lungs andinflammation of the breathing passages.
Zafirlukast is marketed by Astra Zeneca with the brand names Accolate, Accoleit, and Vanticon. It was the first LTRA to be marketed in the USA and is now approved in over 60 countries, including the UK, Japan, Taiwan, Italy, Spain, Canada, Brazil, China and Turkey
Healthy young men who received a single oral 40 mg dose attained peak plasma zafirlukast concentrations that averaged 607 μg/L at 3.4 hours. The elimination half-life ranged from 12 to 20 hours. In another study involving a 20 mg single oral dose in healthy men, the elimination half-life averaged 5.6 hours.[1][2]
A letter was submitted to the FDA by Zeneca Pharmaceuticals on July 22, 1997, notifying them of a change in product labeling that includes the following potential reaction in patients undergoing a dosage reduction of oral steroids who are currently taking zafirlukast:
PRECAUTIONS-Eosinophilic Conditions: The reduction of the oral steroid dose, in some patients on ACCOLATE therapy, has been followed in rare cases by the occurrence of eosinophilia, vasculitic rash, worsening pulmonary symptoms, cardiac complications, and/or neuropathy sometimes presenting as Churg–Strauss syndrome, a systemic eosinophilic vasculitis. Although a causal relationship with ACCOLATE has not been established, caution is required when oral steroid reduction is being considered.1
NDA..020547 26/09/1996, ACCOLATE, ASTRAZENECA, 20MG TABLET
| US Patent No | Expirey Date | patent use code |
|---|---|---|
| 5482963 | Jan 9, 2013 | |
| 5612367 | Mar 18, 2014 | U-189 |
Brief background information
| Salt | ATC | Formula | MM | CAS |
|---|---|---|---|---|
| – | R03DC01 | C 31 H 33 N 3 O 6 S | 575.69 g / mol | 107753-78-6 |
| monohydrate | R03DC01 | C 31 H 33 N 3 O 6 S · H 2 O | 593.70 g / mol | 143052-93-1 |
| calcium (2: 1) | R03DC01 | C 62 H 64 CaN 6 O 12 S 2 | 1189.43 g / mol | 107753-86-6 |
Application
-
antihistamine effect
-
LTD4-antagonist
Classes of substances
-
Benzenesulfonamide (s -imidy), as well as their derivatives
-
Esters of carbamic acid
-
Cyclopentanes
-
Hydroxybenzoic acid amides, and hydroxy acids alkoksibenzoynyh
-
Indoles
-
-
-
-
Zafirlukast is a synthetic, selective peptide leukotriene receptor antagonist (LTRA), with the chemical name 4(5-cyclopentyloxy-carbonylamino-1-methyl-indol-3ylmethyl)-3-methoxy-N-o-tolylsulfonylbenzamide. The molecular weight of zafirlukast is 575.7 and the structural formula is:

Zafirlukast, a fine white to pale yellow amorphous powder, is practically insoluble in water. It is slightly soluble in methanol and freely soluble in tetrahydrofuran, dimethylsulfoxide, and acetone.The empirical formula is: C31H33N3O6S
- Fischer JD, Song MH, Suttle AB, Heizer WD, Burns CB, Vargo DL, Brouwer KL. Comparison of zafirlukast (Accolate) absorption after oral and colonic administration in humans. Pharmaceut. Res. 17: 154-159, 2000.
- Bharathi DV, Naidu A, Jagadeesh B, Laxmi KN, Laxmi PR, Reddy PR, Mullangi R. Development and validation of a sensitive LC-MS/MS method with electrospray ionization for quantitation of zafirlukast, a selective leukotriene antagonist in human plasma: application to a clinical pharmacokinetic study. Biomed. Chromatogr. 22: 645-653, 2008.
- Zafirlukast (U.S. National Library of Medicine)
- Zafirlukast (patient information)

 |
|
 |
|
| Systematic (IUPAC) name | |
|---|---|
| cyclopentyl 3-{2-methoxy-4-[(o-tolylsulfonyl)carbamoyl]benzyl}-1-methyl-1H-indol-5-ylcarbamate | |
| Clinical data | |
| Trade names | Accolate |
| AHFS/Drugs.com | monograph |
| MedlinePlus | a697007 |
| Pregnancy cat. | B1 (Australia), B (United States) |
| Legal status | POM (UK) |
| Routes | Oral |
| Pharmacokinetic data | |
| Bioavailability | Unknown |
| Protein binding | 99% |
| Metabolism | Hepatic (CYP2C9-mediated) |
| Half-life | 10 hours |
| Excretion | Biliary |
| Identifiers | |
| CAS number | 107753-78-6  |
| ATC code | R03DC01 |
| PubChem | CID 5717 |
| IUPHAR ligand | 3322 |
| DrugBank | DB00549 |
| ChemSpider | 5515 |
| UNII | XZ629S5L50 |
| KEGG | D00411 |
| ChEBI | CHEBI:10100 |
| ChEMBL | CHEMBL603 |
| Chemical data | |
| Formula | C31H33N3O6S |
| Mol. mass | 575.676 g/mol |
Trade Names
| Country | Trade name | Manufacturer |
|---|---|---|
| United Kingdom | Akkolat | AstraZeneca |
| Italy | Akkoleit | – “- |
| Zafirst | Chiesi | |
| Japan | Akkolat | AstraZeneca |
| USA | – “- | Zeneca |
| Ukraine | No | No |
Formulations
-
Tablets of 20 mg, 40 mg

is a first anti-asthmatic leukotriene antagonist (Matassa, V.G. et al, J. Med. Chem., v. 33, 1781 ‘(1990); U. S. Patent No. 4,859,692 and The Merck Index, 12th Edition, 10241). Methods for the preparation of Zafirlukast are described in J. Med. Chem., v. 33, 1781 (1990), U. S. Patent 4,859,692 and U.S. Patent 5,993,859 starting from methyl 3-methoxy-4-(l-methyl-5-nitroindol-3-ylmethyl)benzoate [la]


in the presence of an equivalent quantity of silver(I) oxide,

The above process has serious disadvantages in the isolation of the product [4] in step (b) which is due to the fact that alkylation of indole, that is unsubstituted at positions 1-, 2- and 3-, at the 3-position, is accompanied by the undesired process of poly alkylation, to form polysubstituted indoles of formula [7] and/or formula [8] :

while at the same time some quantity of the starting unreacted indole remains in the reaction mixture. Most common methods for the separation of alkyl (indol-3-ylmethyl)benzoate of formula [4] from by-products of polyalkylation and starting unreacted indole, which are all covalent compounds with similar physical properties, include column chromatography that is an unpractical method for industrial scale applications.
Formula (I) compound for the synthesis of an important intermediate of zafirlukast.Reported in the patent EP199543 synthesized compound (I) of the conventional method, the following formula:

(A) (I)
In this method, Intermediate A and 5 – nitro-indole silver oxide in the presence of a catalyst, for docking composite formula (I) compound. Reported only 45% of the reaction yield, the reaction is difficult to complete the reaction and post-treatment using chromatographic methods, resulting in product purification more difficult. And the use of more expensive silver oxide catalysts, high cost.
W00246153 reported a catalyst for the above reaction to zinc bromide, Compound (I), after treatment of the compound (I) with sodium hydroxide hydrolysis of the intermediate (B), separating the product and raw materials purification products.

The method reported in the literature a yield of 60%, but the actual operation is repeated only about 30% yield, and the operation is complicated, cumbersome and costly.
zaafirlukast is a selective and competitive receptor antagonist of leukotriene D4 and E4 (LTD4 and LTE4), components of slow-reacting substance of anaphylaxis (SRSA). Cysteinyl leukotriene production and receptor occupation have been correlated with the pathophysiology of asthma, including airway edema, smooth muscle constriction, and altered cellular activity associated with the inflammatory process, which contribute to the signs and symptoms of asthma.
The cysteinyl leukotrienes (LTC4 LTD4, LTE4) are the products of arachidonic acid metabolism and are various cells, including mast cells and eosinophills, these eicosinoids bind to cysteinyl leukotriene (CysLT) receptors. The CysLT type-1 (CysLT1) receptor is found in human airway and other pro-inflammatory cells. CysLTs have been correlated with the pathophysiology of asthma.
Zafirlukast is a synthetic, selective peptide leukotriene receptor antagonist (LTRA), useful for the treatment of asthma and is commercially available in products sold under the brand name ACCOLATE™ as 10 and 20 mg tablets for oral administration. ACCOLATE™ is indicated for the prophylaxis and treatment of asthma in adults and children 5 years of age and older.
ACCOLATE™ film coated tablets contain amorphous zafirlukast as the active ingredient and the excipients croscarmellose sodium, lactose, magnesium stearate, microcrystalline cellulose, povidone, hypromellose, and titanium dioxide.
The greatest prevalence of asthma is in preschool children; however, the clinical utility of asthma therapy for this age group is limited by a narrow therapeutic index, long-term tolerability, and frequency and/or difficulty of administration. Asthma treatment requires an immediate perceivable effect. Inhalation therapy is a very common therapy prescribed for young children; inhalation therapy has the disadvantage of high dose variability.
 Process for the preparation of zafirlukast
US 20040186300 A1


An Improved and Scalable Process for Zafirlukast: An Asthma Drug
Melting range: 142−145 °C; MS (m/z): 576 (M+ + H); IR (KBr, cm−1): 3326 (NH), 1679 (−C═O), 1H NMR (CDCl3) δ 7.0−8.0 (m, 11H), 3.7 (s, 3H), 4.0 (s, 2H), 3.9 (s, 3H), 2.6 (s, 3H), 1.45−1.8 (s, 9H). ……………………………………………………………….. US 20040186300 A1 http://www.google.com/patents/US20040186300 zafirlukast ethanolate as white powder with mp 132-133° C. (dec.) and 99.8% purity by HPLC. 1H NMR (CDCl3, δ, ppm): 1.22 (t, J 7.05 Hz, 3H), 1.45-1.87 (m, 8H), 2.66 (s, 3H), 3.67 (s, 3H), 3.73 (q, J 7.05 Hz, 4H), 3.79 (s, 3H), 3.98 (s, 2H), 5.08-5.23 (m, 1H), 6.58 (s, 1H), 6.73 (s, 1H), 7.01-7.51 (m, 9H), 8.23 (d, J 7.52 Hz, 1H), 9.67 (s, 1H).
Synthesis pathway
| Synthesis a) |
|---|
      |
| Synthesis of b) |
   |
-
Synthesis a)
-
US 4,859,692 (ICI; 08/22/1989; GB -prior. 4/17/1985; 17.10.1985).
-
EP 199 543 (ICI, Zeneca; appl. 16.4.1986; GB -prior. 4/17/1985).
-
-
Synthesis of b)
-
EP 490 649 (ICI, Zeneca; 11.12.1991; GB -prior. 12.12.1990).
-
Matassa, G. et al .: J. Med. Chem. (JMCMAR) 33, 1781 (1990).
-
Srinivas, K. et al .: Org. Process Res. Dev. (OPRDFK) 8 (6), 952 (2004).
-
added info Asthma is a disease that causes swelling and narrowing the airways of the lungs. Airways are air carriers to and from lungs. Swollen and narrower airways affect the air flow to and from the lungs and this lead to tightness of chest, wheezing, shortness of breath and cough. These symptoms are often occurs in early morning and in night. Asthma is caused by genetic and environmental factors, it was not curable completely but this can be controlled with good medical care. Leukotriene antagonists also known as leukast are the medicaments that are used to reduce leukotrienes, which are produced by several types of cells and causes inflammation in asthma and bronchitis. Leukotriene antagonists that are available in market are Montelukast, Zafirlukast and Pranlukast. Zafirlukast is the first leukast compound approved for management of Asthma. US FDA approved zafirlukast in the form of 10 mg and 20 mg tablet with the brand name of Accolate®.1 Subsequently this was approved and launched by innovator in few other countries. There are many synthetic routes for the preparation of Zafirlukast 4 is well documented in literature. Some of the key approaches are discussed here under. Scientists from ICI Americas Inc2 have reported process for the synthesis of 4, which starts with esterification of 3-methoxy-4-methyl benzoic acid 53 using methanol in presence of acetyl chloride PRODUCT PATENT ROUTE Allylic bromination of methyl ester 54 using bromine in presence of CCl4 resulted bromo compound 55, which was reacted with 5-nitro indole 124 using silver oxide as catalyst to obtain condensed compound 125. N-methylation of 125 utilizing methyl iodide in presence of NaH afforded N-methyl indole derivative 57. Thus obtained 57 was subjected to reduction using palladium carbon (Pd/C) in methanol followed by reacted with cyclopentyl chloroformate to obtain compound 59. Hydrolysis of 59 using LiOH.H2O subsequently reaction with o-toluene sulfonamide (OTSA) in presence of 1-[3-(dimethylamino)propyl]-3-ethyl carbodiimide hydrochloride (DMAPEC) and DMAP furnished zafirlukast 4. Matassa et al3 also reported similar procedure for the synthesis of Zafirlukast 4. 

FDA okays Vifor Fresenius phosphate binder Velphoro
THERAPEUTIC CLAIM Oral phosphate binder, treatement of elevated
phosphate levels in patients undergoing dialysis
CHEMICAL DESCRIPTIONS
1. Ferric hydroxide oxide
2. Mixture of iron(III) oxyhydroxide, sucrose, starches
3. Polynuclear iron(III) oxyhydroxide stabilized with sucrose and starches
structure
O =Fe -OH
MOLECULAR FORMULA FeHO2•xC12H22O11•y(C6H10O5)n
SPONSOR Vifor (International) Inc.
CODE DESIGNATIONS PA21
CAS REGISTRY NUMBER 12134-57-5
sucroferric oxyhydroxide
Sucroferric oxyhydroxide nonproprietary drug name
1. February 27, 2013. N13/36. STATEMENT ON A NONPROPRIETARY NAME ADOPTED BY THE USAN COUNCIL. USAN (ZZ-19). SUCROFERRIC …
The US Food and Drug Administration has given the green light to Vifor Fresenius Medical Care Renal Pharma’s hyperphosphatemia drug Velphoro.
The approval for Velphoro (sucroferric oxyhydroxide), formerly known as PA21, is based on Phase III data demonstrated that the drug successfully controls the accumulation of phosphorus in the blood with the advantage of a much lower pill burden than the current standard of care in patients with chronic kidney disease on dialysis, namely Sanofi’s Renvela (sevelamer carbonate). read this at
http://www.pharmatimes.com/Article/13-11-28/FDA_okays_Vifor_Fresenius_phosphate_binder_Velphoro.aspx
Velphoro (PA21) receives US FDA approval for the treatment of hyperphosphatemia in Chronic Kidney Disease Patients on dialysis
Velphoro (sucroferric oxyhydroxide) has received US Food and Drug Administration (FDA) approval for the control of serum phosphorus levels in patients with Chronic Kidney Disease (CKD) on dialysis. Velphoro will be launched in the US by Fresenius Medical Care North America in 2014.
Velphoro (previously known as PA21) is an iron-based, calcium-free, chewable phosphate binder. US approval was based on a pivotal Phase III study, which met its primary and secondary endpoints. The study demonstrated that Velphoro® successfully controls hyperphosphatemia with fewer pills than sevelamer carbonate, the current standard of care in patients with CKD on dialysis. The average daily dose to control hyperphosphatemia was 3.3 pills per day after 52 weeks.
Velphoro was developed by Vifor Pharma. In 2011, all rights were transferred to Vifor Fresenius Medical Care Renal Pharma, a common company of Galenica and Fresenius Medical Care. In the US, Velphorowill be marketed by Fresenius Medical Care North America, a company with a strong marketing and sales organization, and expertise in dialysis care. The active ingredient of Velphoro is produced by Vifor Pharma in Switzerland.
Hyperphosphatemia, an abnormal elevation of phosphorus levels in the blood, is a common and serious condition in CKD patients on dialysis. Most dialysis patients are treated with phosphate binders. However, despite the availability of a number of different phosphate binders, up to 50% of patients depending on the region are still unable to achieve and maintain their target serum phosphorus levels. In some patients, noncompliance due to the high pill burden and poor tolerability appear to be key factors in the lack of control of serum phosphorus levels. On average, dialysis patients take approximately 19 pills per day with phosphate binders comprising approximately 50% of the total daily pill burden. The recommended starting dose of Velphoro is 3 tablets per day (1 tablet per meal).
Full results from the pivotal Phase III study involving more than 1,000 patients were presented at both the 50th ERA-EDTA (European Renal Association European Dialysis and Transplant Association) Congress in Istanbul, Turkey, in May 2013, and the American Society of Nephrology (ASN) Kidney Week in Atlanta, Georgia, in November 2013. Velphorowas shown to be a potent phosphate binder, with lower pill burden and a good safety profile.
Based on these data, Vifor Fresenius Medical Care Renal Pharma believes that Velphoro offers a new and effective therapeutic option for the control of serum phosphorus levels in patients with chronic kidney disease on dialysis.
The regulatory processes in Europe, Switzerland and Singapore are ongoing and decisions are expected in the first half 2014. Further submissions for approval are being prepared.
Teva Gets Orphan Drug Designation for Treanda
Teva Announces Additional Regulatory Exclusivity for TREANDA® (Bendamustine HCI) for Injection
Orphan Designation combined with pediatric extension provides regulatory exclusivity through April 2016 for indolent B-cell non-Hodgkin lymphoma indication
JERUSALEM, November 27, 2013 –(BUSINESS WIRE)–Teva Pharmaceutical Industries Ltd. (NYSE: TEVA) today announced that the U.S. Food and Drug Administration (FDA) has granted orphan drug exclusivity for TREANDA through October 2015 for indolent B-cell non-Hodgkin lymphoma (iNHL) that has progressed during or within six months of treatment with rituximab or a rituximab-containing regimen.http://www.pharmalive.com/teva-announces-additional-regulatory-exclusivity-for-treanda
read my old post, contains synthesis
https://newdrugapprovals.wordpress.com/2013/09/19/fda-oks-tevas-injectable-treanda/
VBL Therapeutics announced FDA has granted Fast Track designation to its lead oncology drug VB-111

VB-111
VBL Therapeutics announced today that the U.S. Food and Drug Administration (FDA) has granted Fast Track designation to its lead oncology drug VB-111, for prolongation of survival in patients with recurrent glioblastoma multiforme (rGBM).
VB-111 – highly targeted anti-angiogenic agent for the specific inhibition of tumor vascular growth
VB-111 is the first highly targeted anti-angiogenic agent for the specific inhibition of tumor vascular growth to use VTS™™, our proprietary platform technology, for cancer therapy. VB-111 is an IV-administered anti angiogenic agent that works in a manner akin to a “biological knife” to destroy tumor vasculature, thus cutting off blood vessels feeding the tumor.
Preclinical Insights
VB-111 has shown significant promise as a targeted cancer treatment with the potential to work synergistically in combination with conventional chemotherapy treatments to provide an effective treatment regimen for cancer patients. Pharmacological and toxicology studies of VB-111 have showed tissue specificity for the tumor tissue, no significant damage to normal non-cancerous tissues or to the normal vasculatures in the body and more than 90 percent tumor burden reduction in a metastatic lung cancer model with only one injection. Similar efficacy was shown in other tumor models.
Completed Clinical Trials
Phase 1 Clinical Trial – in a Phase 1 “all comers” dose escalation study in 33 patients with advanced metastatic cancer, therapeutic doses of VB-111 demonstrated antitumor activity and was found to be safe and well tolerated with no effect on liver function or major changes in complete blood count. Findings have been presented at the American Association of Cancer Research (AACR) and the American Society of Clinical Oncology (ASCO) annual meetings.
GSK obtains FDA approval for bird flu vaccine
GlaxoSmithKline (GSK) has received approval from the US Food and Drug Administration (FDA) for the first adjuvanted vaccine to prevent H5N1 influenza, also known as bird flu.
GSK obtains FDA approval for bird flu vaccine http://www.pharmaceutical-technology.com/news/newsgsk-obtains-fda-approval-bird-flu-vaccine?WT.mc_id=DN_News
26 November 2013

GlaxoSmithKline (GSK) has received approval from the US Food and Drug Administration (FDA) for the first adjuvanted vaccine to prevent H5N1 influenza, also known as bird flu.
The FDA cleared the pandemic Influenza A (H5N1) virus monovalent vaccine, adjuvanted (also referred to as Q-Pan H5N1 influenza vaccine), for use in people aged 18 and older who are at increased risk of exposure to the virus.
The vaccine is composed of monovalent, inactivated, split A/H5N1 influenza virus antigen and GSK’s AS03 adjuvant.
The company said that in clinical studies, the adjuvanted formulation stimulated the required immune response while using a smaller amount of antigen as compared with a formulation without adjuvant.
FDA Approves Olysio (simeprevir) for Hepatitis C Virus

Simeprevir
Inhibits HCV NS3/4A protease.
MEDIVIR … originator
launched 2013
923604-59-5 CAS
C38H47N5O7S MF
749.93908 MW
IUPAC standard name
(1R, 4R, 6S, 15R, 17R)-N-(cyclopropanesulfonyl) -17 – ({7-methoxy-8-methyl-2-[4 – (propan-2-yl) -1,3-thiazol-2 -yl] quinolin-4-yl} oxy)-13-methyl-2 ,14-dioxo-3 ,13-diazatricyclo [13.3.0.0 4 , 6 ] octadec-7-ene-4-carboxamide
IUPAC traditional name
(1R, 4R, 6S, 15R, 17R)-N-(cyclopropanesulfonyl) -17 – {[2 – (4-isopropyl-1 ,3-thiazol-2-yl)-7-methoxy-8-methylquinolin-4- yl] oxy}-13-methyl-2 ,14-dioxo-3 ,13-diazatricyclo [13.3.0.0 4 , 6 ] octadec-7-ene-4-carboxamide
- Olysio
- Simeprevir
- TMC 435
- TMC 435350
- TMC-435
- TMC435
- TMC435350
- UNII-9WS5RD66HZ
November 22, 2013 — The U.S. Food and Drug Administration approved Olysio (simeprevir), a new therapy to treat chronic hepatitis C virus infection.
OLYSIO™ is the first once-daily protease inhibitor approved for the treatment of chronic hepatitis C in a combination antiviral regimen for adults with compensated liver disease
Hepatitis C is a viral disease that causes inflammation of the liver that can lead to diminished liver function or liver failure. Most people infected with the hepatitis C virus have no symptoms of the disease until liver damage becomes apparent, which may take several years. Most of these people then go on to develop chronic hepatitis C. Some will also develop scarring and poor liver function (cirrhosis) over many years, which can lead to complications such as bleeding, jaundice (yellowish eyes or skin), fluid accumulation in the abdomen, infections or liver cancer. According to the Centers for Disease Control and Prevention, about 3.2 million Americans are infected with the hepatitis C virus

Hepatitis C virus (HCV) infections affect approximately 3 percent of the worldwide population and often lead to cirrhosis and hepatocellular carcinoma. The standard therapy of pegylated- interferon and ribavirin induces serious side effects and provides viral eradication in less than 50% of patients. Combination therapy of HCV including ribavirin and interferonare currently is the approved therapy for HCV. Unfortunately, such combination therapy also produces side effects and is often poorly tolerated, resulting in major clinical challenges in a significant proportion of patients. Numerous direct acting agents (DAAs) have been or are being developed for treatment of HCV, such as telaprevir and boceprevir (both received MA approved in 2011 for use with interferon and ribavirin based therapy), however direct acting agents are linked to increased toxicity of treatment, the emergence of resistance, and to date do not provide a standard of care which is interferon free. The combination of direct acting agents can also result in drug-drug interactions. To date, no HCV therapy has been approved which is interferon free. There is therefore a need for new combination therapies which have reduced side effects, and interferon free, have a reduced emergence of resistance, reduced treatment periods and/or and enhanced cure rates.
Simeprevir (formerly TMC435) is an experimental drug candidate for the treatment of hepatitis C. It is being developed byMedivir and Johnson & Johnson‘s pharmaceutical division Janssen Pharmaceutica and is currently in Phase III clinical trials.[1]
Simeprevir is a hepatitis C virus protease inhibitor.[2]
Simeprevir is being tested in combination regimens with pegylated interferon alfa-2a and ribavirin,[3] and in interferon-free regimens with other direct-acting antiviral agents including daclatasvir[4] and sofosbuvir [5]
Simeprevir has been launched in 2013 in Japan by Janssen Pharmaceutical (JP) for use in combination with pegylated interferon (Peg-IFN) and ribavirin for the treatment of genotype 1 chronic hepatitis C virus (HCV) patients who are treatment naïve, prior non responders or relapsed following treatment with Peg-IFN with or without ribavirin. In 2013, the product has also been approved in the U.S. by Medivir and Janssen R&D Ireland for the oral treatment of chronic hepatitis C genotype 1 infection, in combination with peginterferon alfa and ribavirin in adults with compensated liver disease, including cirrhosis, who are treatment-naïve or who have failed previous interferon therapy (pegylated or non-pegylated) with ribavirin.
The drug candidate was originally developed at Medivir, which was acquired by Janssen R&D Ireland in 2012. In November 2004, Medivir entered into a license and research collaboration agreement with Tibotec, a Johnson & Johnson subsidiary, for the discovery and development of orally active protease inhibitors of the NS3/4A protease of HCV. In 2011, a codevelopment agreement between Pharmasset (now Gilead Sciences) and Tibotec was signed for the treatment of chronic hepatitis C (HCV) in combination with PSI-7977. Also in 2011, fast track designation was received in the U.S. for the treatment of chronic hepatitis C (CHC) genotype-1 infection.
In 2011, Tibotec Therapeutics, Division of Centocor Ortho Biotech Products, L.P. announced that it had changed its name to Janssen Therapeutics, Division of Janssen Products, LP.
“Hepatitis C is a complex disease and Janssen is committed to working with the HCV community, caregivers, and health care systems to address this global epidemic,” said Gaston Picchio, Hepatitis Disease Area Leader, Janssen Research & Development. “We are pleased that the FDA has granted simeprevir Priority Review, as it is a significant step forward in making this therapy available to physicians and their hepatitis C patients.”

Hepatitis C virus (HCV) is the leading cause of chronic liver disease worldwide.
Following initial acute infection, a majority of infected individuals develop chronic hepatitis because HCV replicates preferentially in hepatocytes but is not directly cytopathic. Chronic hepatitis can progress to liver fibrosis leading to cirrhosis, end- stage liver disease, and HCC (hepatocellular carcinoma), making it the leading cause of liver transplantations. This and the number of patients involved, has made HCV the focus of considerable medical research. Replication of the genome of HCV is mediated by a number of enzymes, amongst which is HCV NS3 serine protease and its associated cofactor, NS4A. NS3 serine protease is considered to be essential for viral replication and has become an attractive target for drug discovery.
Current anti-HCV therapy is based on (pegylated) interferon-alpha (IFN-α) in combination with ribavirin. Not only does this therapy result in a limited efficacy in that only part of the patients are treated successfully, but it also faces significant side effects and is poorly tolerated in many patients. Hence there is a need for further HCV inhibitors that overcome the disadvantages of current HCV therapy such as side effects, limited efficacy, poor tolerance, the emergence of resistance, as well as compliance failures.
Various agents have been described that inhibit HCV NS3 serine protease. WO05/073195 discloses linear and macrocyclic NS3 serine protease inhibitors with a central substituted proline moiety and WO 05/073216 with a central cyclopentyl moiety. Amongst these, the macrocyclic derivatives are attractive by overcoming one or more of the disadvantages of current anti-HCV therapy

(I) simeprevir
The compound of formula (I) is an inhibitor of the Hepatitis C virus (HCV) serine protease and is described in WO 2007/014926, published on 8 February 2007. This compound overcomes several of the disadvantages of current anti-HCV therapy and in particular shows pronounced activity against HCV, has an attractive pharmacokinetic profile, and is well-tolerated. Following the synthesis procedure described in Example 5 of WO 2007/014926, an amorphous solid form is obtained.
It now has been found that the compound of formula (I) can be converted into crystalline forms, which can advantageously be used as active ingredients in anti-HCV therapy. To that purpose, these crystalline forms are converted into pharmaceutical formulations.
………………………………………………………………………………………….
SIMEPREVIR

…………………………
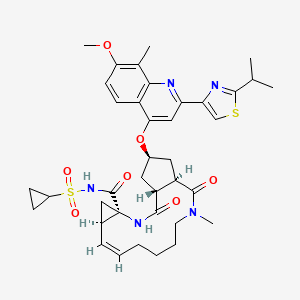
OLYSIO (simeprevir) is an inhibitor of the HCV NS3/4A protease.
The chemical name for simeprevir is (2R,3aR,10Z,11aS,12aR,14aR)-N-(cyclopropylsulfonyl)-2[[2-(4-isopropyl-1,3-thiazol-2-yl)-7-methoxy-8-methyl-4-quinolinyl]oxy]-5-methyl-4,14-dioxo2,3,3a,4,5,6,7,8,9,11a,12,13,14,14atetradecahydrocyclopenta[c]cyclopropa[g][1,6]diazacyclotetradecine-12a(1H)-carboxamide. Its molecular formula is C38H47N5O7S2 and its molecular weight is 749.94. Simeprevir has the following structural formula:
 |
Simeprevir drug substance is a white to almost white powder. Simeprevir is practically insoluble in water over a wide pH range. It is practically insoluble in propylene glycol, very slightly soluble in ethanol, and slightly soluble inacetone. It is soluble in dichloromethane and freely soluble in some organic solvents (e.g., tetrahydrofuran and N,N-dimethylformamide).
OLYSIO (simeprevir) for oral administration is available as 150 mg strength hard gelatin capsules. Each capsule contains 154.4 mg of simeprevir sodium salt, which is equivalent to 150 mg of simeprevir. OLYSIO (simeprevir) capsules contain the following inactive ingredients: colloidal anhydrous silica, croscarmellose sodium, lactose monohydrate, magnesium stearate and sodium lauryl sulphate. The white capsule contains gelatin and titanium dioxide (E171) and is printed with ink containing iron oxide black (E172) and shellac (E904).
……………..
Synthesis
Example 1 : preparation of 17-[2-(4-isopropylthiazole-2-yl)-7-methoxy-8-methyl- quinolin-4-yloxy]- 13-methyl-2, 14-dioxo-3, 13-diazatricyclo[ 13.3.0.04‘6]octadec-7-ene- 4-carboxylic acid (16)
Synthesis of 4-hydroxy-2-(4-isopropylthiazole-2-yl)-7-methoxy-8-methylquinoline (6) Step 1 : synthesis of Λ/-(tert-butyloxycarbonyl)-3-methoxy-2-methylaniline (2)

1 2
Triethylamine (42.4 mL, 302 mmol) was added to a suspension of 3-methoxy-2- methylbenzoic acid (45.6 g, 274 mmol) in dry toluene (800 mL). A clear solution was obtained. Then, dppa (65.4 mL, 302 mmol) in toluene (100 mL) was slowly added. After 1 h at room temperature, the reaction mixture was successively heated at 500C for 0.5 h, at 700C for 0.5 h then at 1000C for 1 h. To this solution, t-BuOH (30.5 g, 411 mmol) in toluene (40 mL) was added at 1000C and the resulting mixture was refluxed for 7h. The solution was cooled to room temperature then successively washed with water, 0.5 N HCl, 0.5 N NaOH and brine, dried (Na2SO4), and evaporated to give 67 g of the target product: m/z = 237 (M)+.
_2: synthesis of 3-methoxy-2-methylaniline (3)

TFA (40.7 mL, 548 mmol) was added to a solution of jV-(teτt-butyloxycarbonyl)- 3-methoxy-2-methylaniline, in dichloro methane (500 mL). After 2 h at room temperature, TFA (40.7 mL, 548 mmol) was added and the resulting mixture was stirred at room temperature overnight. Then, volatiles were evaporated. The residue was triturated with toluene (100 mL) and diisopropylether (250 mL), filtered off and washed with diisopropyl ether (100 mL) to give 56.3 g of the title product as a TFA salt: m/z = 138 (M+H)+. The TFA salt was transformed to the free aniline by treatment with NaHCO3.
Step 3: synthesis of (2-amino-4-methoxy-3-methylphenyl)(methyl)ketone (4)

A solution Of BCl3 (1.0 M, 200 mL, 200 mmol) in CH2Cl2 was slowly added under nitrogen to a solution of 3-methoxy-2-methylaniline (26.0 g, 190 mmol) in xylene (400 mL). The temperature was monitored during the addition and was kept below 100C. The reaction mixture was stirred at 5°C for 0.5 h. Then, dry acetonitrile (13 mL, 246 mmol) was added at 5°C. After 0.5 h at 5°C, the solution was transferred into a dropping funnel and slowly added at 5°C to a suspension OfAlCl3 (26.7 g, 200 mmol) in CH2Cl2 (150 mL). After 45 min at 5°C, the reaction mixture was heated at 700C under a nitrogen stream. After evaporation Of CH2Cl2, the temperature of the reaction mixture reached 65°C. After 12 h at 65°C, the reaction mixture was cooled at 00C, poured onto ice (300 g), and slowly heated to reflux for 7h. After 2 days at room temperature, 6 N NaOH (50 mL) was added. The pH of the resulting solution was 2-3. The xylene layer was decanted. The organic layer was extracted with CH2Cl2. The xylene and CH2Cl2 layers were combined, successively washed with water, IN NaOH, and brine, dried (Na2SO4) and evaporated. The residue was triturated in diisopropyl ether at O0C, filtered off and washed with diisopropylether to give 13.6 g (40 %) of the title product as a yellowish solid: m/z = 180 (M+H)+.
Step 4: synthesis of 2′-[[(4-isopropylthiazole-2-yl)(oxo)methyl]amino]-4′-methoxy-3 ‘- methylacetophenone (5)

A solution of the compound 4 (18.6 g, 104 mmol) in dioxane (50 rnL) was added under nitrogen to a suspension of 4-isopropylthiazole-2-carbonyl chloride in dioxane (250 rnL). After 2 h at room temperature, the reaction mixture was concentrated to dryness. Then, the residue was partitioned between an aqueous solution of NaHCOs and AcOEt, organic layer was washed with brine, dried (Na2SO4), and evaporated. The residue was triturated in diisopropyl ether, filtered off and washed with diisopropyl ether to give 30.8 g (90 %) of the title product 5.
Step 5: synthesis of 4-hydroxy-2-(4-isopropylthiazole-2-yl)-7-methoxy-8- methylquinoline (6)

Potassium tert-butoxide (21.8 g, 195 mmol) was added to a suspension of the compound 5 (30.8 g, 92.7 mmol) in tert-butanol. The resulting reaction mixtures was heated at 1000C overnight. Then, the reaction mixture was cooled at room temperature and diluted with ether (100 mL). The precipitate was filtered off and washed with Et2O to give a powder (fraction A). The mother liquor was concentrated in vacuo, triturated in ether, filtered off, and washed with ether to give a powder (fraction 2). Fractions 1 and 2 were mixed and poured into water (250 mL). The pH of the resulting solution was adjusted to 6-7 (control with pH paper) with HCl IN. The precipitate was filtered off, washed with water and dried. Then, the solid was triturated in diisopropyl ether, fϊltered off and dried to give 26 g (88%) of the compound 6 as a brownish solid: m/z = 315 (M+H)+.
Synthesis of (hex-5-enyl)(methyl)amine (8)
O CF,
FX N Br’ N O NH
H 7
(a) Sodium hydride (1.05 eq) was slowly added at 00C to a solution of JV-methyl- trifluoro-acetamide (25 g) in DMF (140 mL). The mixture was stirred for Ih at room temperature under nitrogen. Then, a solution of bromohexene (32,1 g) in DMF
(25 mL) was added dropwise and the mixture was heated to 700C for 12 hours. The reaction mixture was poured on water (200 mL) and extracted with ether (4 x 50 mL), dried (MgSO4), filtered and evaporated to give 35 g of the target product 7 as a yellowish oil which was used without further purification in the next step.
(b) A solution of KOH (187.7 g) in water (130 mL) was added dropwise to a solution of 7 (35 g) in methanol (200 mL). The mixture was stirred at room temperature for
12 hours. Then, the reaction mixture was poured on water (100 mL) and extracted with ether (4 x 50 mL), dried (MgSO4), filtered and the ether was distilled under atmospheric pressure. The resulting oil was purified by distillation under vacuum (13 mm Hg pressure, 500C) to give 7,4 g (34 %) of the title product 8 as a colourless oil: 1H-NMR (CDCl3): δ 5.8 (m, IH), 5 (ddd, J = Yl 2 Hz, 3.5 Hz, 1.8 Hz, IH), 4.95 (m, IH), 2.5 (t, J = 7.0 Hz, 2H), 2.43 (s, 3H), 2.08 (q, J= 7.0 Hz, 2H), 1.4 (m, 4H), 1.3 (br s, IH).
Preparation of 17-[2-(4-isopropylthiazole-2-yl)-7-methoxy-8-methylquinolin-4-yloxyl- 13-methyl-2, 14-dioxo-3, 13-diazatricyclo[ 13.3.0.04‘6loctadec-7-ene-4-carboxylic acid (16)

3-Oxo-2-oxa-bicyclo[2.2.1]heptane-5-carboxylic acid 9 (500 mg, 3.2 mmol) in 4 mL DMF was added at 00C to HATU (1.34 g, 3.52 mmol) and JV-methylhex-5-enylamine (435 mg, 3.84 mmol) in DMF (3 mL), followed by DIPEA. After stirring for 40 min at 00C, the mixture was stirred at room temperature for 5 h. Then, the solvent was evaporated, the residue dissolved in EtOAc (70 rnL) and washed with saturated NaHCOs (IO mL). The aqueous layer was extracted with EtOAc (2 x 25 mL). The organic phases were combined, washed with saturated NaCl (20 mL), dried (Na2SO4), and evaporated. Purification by flash chromatography (EtO Ac/petroleum ether, 2:1) afforded 550 mg (68%) of the target product 10 as a colorless oil: m/z = 252 (M+H)+.

A solution of LiOH (105 mg in 4 mlof water) was added at 00C to the lactone amide 10. After Ih, the conversion was completed (HPLC). The mixture was acidified to pH 2 – 3 with IN HCl, extracted with AcOEt, dried (MgSO4), evaporated, co-evaporated with toluene several times, and dried under high vacuum overnight to give 520 mg (88%) of the target product 11: m/z = 270 (M+H)+.

The l-(amino)-2-(vinyl)cyclopropanecarboxylic acid ethyl ester hydrochloride 12
(4.92 g, 31.7 mmol) and HATU (12.6 g, 33.2 mmol) were added to 11 (8.14 g,
30.2 mmol). The mixture was cooled in an ice bath under argon, and then DMF (100 mL) and DIPEA (12.5 mL, 11.5 mmol) were successively added. After 30 min at 00C, the solution was stirred at room temperature for an additional 3 h. Then, the reaction mixture was partitioned between EtOAc and water, washed successively with 0.5 N HCl (20 mL) and saturated NaCl (2 x 20 mL), and dried (Na2SO4). Purification by flash chromatography (AcOEt/CH2Cl2/Petroleum ether, 1 :1 :1) afforded 7.41 g (60%) of the target product 13 as a colorless oil: m/z = 407 (M+H)+.

DIAD (1.02 niL, 5.17 mmol) was added at -15°C under nitrogen atmosphere to a solution of 13 (1.5 g, 3.69 mmol), quinoline 6 (1.39 g, 4.43 mmol) and triphenyl- phosphine (1.26 g, 4.80 mmol) in dry THF (40 mL). After 4.5 h, at -15°C, the reaction mixture was partitioned between ice-cold water and AcOEt, dried (Na2SO4) and evaporated. The crude material was purified by flash column chromatography (gradient of petroleum AcOEt/CH2Cl2, 1 :9 to 2:8) to give 1.45 g (56 %) of the target product 14: m/z = 703 (M+H)+.

A solution of 14 (1.07 g, 1.524 mmol) and Hoveyda-Grubbs 1st generation catalyst (33 mg, 0.03 eq) in dried and degassed 1 ,2-dichloroethane (900 mL) was heated at 75°C under nitrogen for 12 h. Then, the solvent was evaporated and the residue purified by silica gel chromatography (25% EtOAc in CH2Cl2). 620 mg (60%) of pure macrocycle 15 were obtained, m/z = 674 (M+H)+. 1H NMR (CDCl3): 1.18-1.39 (m, 12H), 1.59 (m, IH), 1.70-2.08 (m, 5H), 2.28 (m, IH), 2.38 (m, IH), 2.62 (m, 2H), 2.68 (s, 3H), 2.83 (m, IH), 3.06 (s, 3H), 3.19 (sept, J= 6.7 Hz, IH), 3.36 (m, IH), 3.83 (m, IH), 3.97 (s, 3H), 4.09 (m, 2H), 4.65 (td, J= 4 Hz, 14 Hz, IH), 5.19 (dd, J= 4 Hz,
10 Hz, IH), 5.31 (m, IH), 5.65 (td, J= 4 Hz, 8 Hz, IH), 7.00 (s, IH), 7.18 (s, IH), 7.46
(d, J= 9 Hz, IH), 7.48 (s, IH), 8.03 (d, J= 9 Hz, IH).

A solution of lithium hydroxide (1.65 g, 38.53 mmol) in water (15 rnL) was added to a stirred solution of ester 15 (620 mg, 0.920 mmol) in THF (30 mL) and MeOH (20 mL). After 16 h at room temperature, the reaction mixture was quenched with NH4Cl sat., concentrated under reduced pressure, acidified to pH 3 with HCl IN and extracted with CH2Cl2, dried (MgSO4) and evaporated to give 560 mg (88%) of carboxylic acid 16. m/z = 647 (M+H)+. 1H NMR (CDCl3): 1.11-1.40 (m, 8H), 1.42-1.57 (m, 2H), 1.74 (m, 2H), 1.88-2.00 (m, 2H), 2.13 (m, IH), 2.28 (m, IH), 2.40 (m, IH), 2.59 (m, 2H), 2.67 (s, 3H), 2.81 (m, IH), 2.97 (s, 3H), 3.19 (m, IH), 3.31 (m, IH), 3.71 (m, IH), 3.96 (s, 3H), 4.56 (dt, J= 4 Hz, 12 Hz, IH), 5.23 (m, 2H), 5.66 (m, IH), 7.01 (s, IH), 7.10 (s, IH), 7.22 (d, J= IO Hz, IH), 7.45 (s, IH), 8.00 (d, J= 10 Hz, IH).
Example 2: Preparation of Λ/-[17-[2-(4-isopropylthiazole-2-yl)-7-methoxy-8-methyl- quinolin-4-yloxy]- 13-methyl-2, 14-dioxo-3, 13-diazatricyclo[ 13.3.0.04‘6]octadec-7-ene- 4-carbonyll(cvclopropyl)sulfonamide (17) SIMEPREVIR

A solution of the compound 16 (560mg, 0.867 mmol) prepared according to Example 4, and carbonyldiimidazole (308 mg, 1.90 mmol) in dry THF (10 mL) was stirred at reflux under nitrogen for 2h. The reaction mixture was cooled to room temperature and cyclopropylsulfonamide (400 mg, 3.301 mmol) and DBU (286 mg, 1.881 mmol) were added. This solution was heated at 500C for 15 h. Then, the reaction mixture was cooled down at room temperature and concentrated under reduced pressure. The residue was partitioned between CH2Cl2 and HCl 1 N, the organic layer was washed with brine, dried (MgSO4) and evaporated. Purification by flash chromatography (gradient of EtOAc (0 to 25%) in CH2Cl2) afforded 314 mg of an off-white solid which was further washed with water, then isopropylether, and dried in the vacuum oven to deliver 282 mg (40%) of the pure title product 17, which is the compound of formula (I) SIMEPREVIR , as a white powder: m/z = 750 (M+H)+.
1H NMR (CDCl3): 0.99-1.52 (m, 14H), 1.64-2.05 (m, 4H), 2.77 (m, IH), 2.41 (m, 2H), 2.59 (m, 2H), 2.69 (s, 3H), 2.92 (m, 2H), 3.04 (s, 3H), 3.19 (m, IH), 3.40 (m, 2H), 3.98 (s, 3H), 4.60 (t, J= 13 Hz, IH), 5.04 (t, J= 11 Hz, IH), 5.37 (m, IH), 5.66 (m, IH), 6.21 (s, IH), 7.02 (s, IH), 7.22 (d, J= IO Hz, IH), 7.45 (s, IH), 7.99 (d, J= 10 Hz, IH), 10.82 (broad s, IH).
…………………
SYNTHESIS
Example 4: preparation of 17-[2-(4-isopropylthiazole-2-yl)-7-methoxy-8-methyl- quinolin-4-yloxy] – 13 -methyl-2, 14-dioxo-3 , 13 -diazatricyclo[ 13.3.0.04‘6]octadec-7-ene- 4-carboxylic acid (46) FREE ACID
Synthesis of 4-hvdroxy-2-(4-isopropylthiazole-2-yl)-7-methoxy-8-methylquinoline (36) Step 1: synthesis of iV-(tert-butyloxycarbonyl)-3-methoxy-2-methylaniline (32)

31 32
Triethylamine (42.4 mL, 302 mmol) was added to a suspension of 3-methoxy-2- methylbenzoic acid (45.6 g, 274 mmol) in dry toluene (800 mL). A clear solution was obtained. Then, dppa (65.4 mL, 302 mmol) in toluene (100 mL) was slowly added. After 1 h at room temperature, the reaction mixture was successively heated at 50°C for 0.5 h, at 70°C for 0.5 h then at 100°C for 1 h. To this solution, t-BuOH (30.5 g, 411 mmol) in toluene (40 mL) was added at 100°C and the resulting mixture was refluxed for 7h. The solution was cooled to room temperature then successively washed with water, 0.5 N HCl, 0.5 N NaOH and brine, dried (Na2SO4), and evaporated to give 67 g of the target product: m/z = 237 (M)+.
Step 2: synthesis of 3-methoxy-2-methylaniline (33)

TFA (40.7 mL, 548 mmol) was added to a solution of iV-(tert-butyloxycarbonyl)-3- methoxy-2-methylaniline, in dichloromethane (500 mL). After 2 h at room temperature, TFA (40.7 mL, 548 mmol) was added and the resulting mixture was stirred at room temperature overnight. Then, volatiles were evaporated. The residue was triturated with toluene (100 mL) and diisopropylether (250 mL), filtered off and washed with diisopropyl ether (100 mL) to give 56.3 g of the title product as a TFA salt: m/z = 138 (M+H)+. The TFA salt was transformed to the free aniline by treatment with NaHCO3.
Step 3: synthesis of (2-amino-4-methoxy-3-methylphenyl)(methyl)ketone (34)

A solution OfBCl3 (1.0 M, 200 mL, 200 mmol) in CH2Cl2 was slowly added under nitrogen to a solution of 3-methoxy-2-methylaniline (26.0 g, 190 mmol) in xylene (400 mL). The temperature was monitored during the addition and was kept below 10°C. The reaction mixture was stirred at 5°C for 0.5 h. Then, dry acetonitrile (13 mL, 246 mmol) was added at 5°C. After 0.5 h at 5°C, the solution was transferred into a dropping funnel and slowly added at 5°C to a suspension OfAlCl3 (26.7 g, 200 mmol) in CH2Cl2 (150 mL). After 45 min at 5°C, the reaction mixture was heated at 70°C under a nitrogen stream. After evaporation Of CH2Cl2, the temperature of the reaction mixture reached 65°C. After 12 h at 65°C, the reaction mixture was cooled at 0°C, poured onto ice (300 g), and slowly heated to reflux for 7h. After 2 days at room temperature, 6 N NaOH (50 mL) was added. The pH of the resulting solution was 2-3. The xylene layer was decanted. The organic layer was extracted with CH2Cl2. The xylene and CH2Cl2 layers were combined, successively washed with water, IN NaOH, and brine, dried (Na2SO4) and evaporated. The residue was triturated in diisopropyl ether at O0C, filtered off and washed with diisopropylether to give 13.6 g (40 %) of the title product as a yellowish solid: m/z = 180 (M+H)+.
Step 4: synthesis of 2′-[[(4-isopropylthiazole-2-yl)(oxo)methyl]amino]-4′-methoxy-3 ‘- methylacetophenone (35)

A solution of (2-amino-4-methoxy-3-methylphenyl)(methyl)ketone (18.6 g, 104 mmol) in dioxane (50 mL) was added under nitrogen to a suspension of 4-isopropylthiazole-2- carbonyl chloride in dioxane (250 mL). After 2 h at room temperature, the reaction mixture was concentrated to dryness. Then, the residue was partitioned between an aqueous solution OfNaHCO3and AcOEt, organic layer was washed with brine, dried (Na2SO4), and evaporated. The residue was triturated in diisopropyl ether, filtered off and washed with diisopropyl ether to give 30.8 g (90 %) of the title product 35.
Step 5: synthesis of 4-hydroxy-2-(4-isopropylthiazole-2-yl)-7-methoxy-8- methylquinoline (36)

Potassium tert-butoxide (21.8 g, 195 mmol) was added to a suspension of 2′-[[(4-iso- propylthiazole-2-yl)(oxo)methyl]amino]-4′-methoxy-3′-methylacetophenone (35, 30.8 g, 92.7 mmol) in tert-butanol. The resulting reaction mixtures was heated at 100°C overnight. Then, the reaction mixture was cooled at room temperature and diluted with ether (100 mL). The precipitate was filtered off and washed with Et2O to give a powder (fraction A). The mother liquor was concentrated in vacuo, triturated in ether, filtered off, and washed with ether to give a powder (fraction 2). Fractions 1 and 2 were mixed and poured into water (250 mL). The pH of the resulting solution was adjusted to 6-7 (control with pH paper) with HCl IN. The precipitate was filtered off, washed with water and dried. Then, the solid was triturated in diisopropyl ether, filtered off and dried to give 26 g (88%) of the title product 36 as a brownish solid: m/z = 315 (M+H)+.
Synthesis of (hex-5-enyl)(methyl)amine (38)

Sodium hydride (1.05 eq) was slowly added at 0°C to a solution of iV-methyltrifluoro- acetamide (25 g) in DMF (140 mL). The mixture was stirred for Ih at room temperature under nitrogen. Then, a solution of bromohexene (32,1 g) in DMF (25 mL) was added dropwise and the mixture was heated to 70°C for 12 hours. The reaction mixture was poured on water (200 mL) and extracted with ether (4 x 50 mL), dried (MgSO4), filtered and evaporated to give 35 g of the target product 37 as a yellowish oil which was used without further purification in the next step.
Step B:
A solution of potassium hydroxide (187.7 g) in water (130 mL) was added dropwise to a solution of 37 (35 g) in methanol (200 mL). The mixture was stirred at room temperature for 12 hours. Then, the reaction mixture was poured on water (100 mL) and extracted with ether (4 x 50 mL), dried (MgSO4), filtered and the ether was distilled under atmospheric pressure. The resulting oil was purified by distillation under vacuum (13 mm Hg pressure, 50°C) to give 7,4 g (34 %) of the title product 38 as a colourless oil: 1H-NMR (CDCl3): δ 5.8 (m, IH), 5 (ddd, J= 17.2 Hz, 3.5 Hz, 1.8 Hz, IH), 4.95 (m, IH), 2.5 (t, J= 7.0 Hz, 2H), 2.43 (s, 3H), 2.08 (q, J= 7.0 Hz, 2H), 1.4 (m, 4H), 1.3 (br s, IH).
Preparation of 17-r2-(4-isopropylthiazole-2-yl)-7-methoxy-8-methylquinolin-4-yloxyl-
13-methyl-2,14-dioxo-3,13-diazatricvclori3.3.0.04‘6loctadec-7-ene-4-carboxylic acid
£46}

3-Oxo-2-oxa-bicyclo[2.2.1]heptane-5-carboxylic acid 39 (500 mg, 3.2 mmol) in 4 mlDMF was added at 0°C to HATU (1.34 g, 3.52 mmol) and iV-methylhex-5- enylamine (435 mg, 3.84 mmol) in DMF (3 mL), followed by DIPEA. After stirring for 40 min at 0°C, the mixture was stirred at room temperature for 5 h. Then, the solvent was evaporated, the residue dissolved in EtOAc (70 mL) and washed with saturated NaHCO3 (10 mL). The aqueous layer was extracted with EtOAc (2 x 25 mL). The organic phases were combined, washed with saturated NaCl (20 mL), dried (Na2SO4), and evaporated. Purification by flash chromatography (EtOAc/petroleum ether, 2:1) afforded 550 mg (68%) of the target product 40 as a colorless oil: m/z = 252 (M+H)+.

A solution of LiOH (105 mg in 4 mlof water) was added at 0°C to the lactone amide 40. After Ih, the conversion was completed (HPLC). The mixture was acidified to pH 2 – 3 with IN HCl, extracted with AcOEt, dried (MgSO4), evaporated, co-evaporated with toluene several times, and dried under high vacuum overnight to give 520 mg (88%) of the target product 41: m/z = 270 (M+H)+.

The l-(amino)-2-(vinyl)cyclopropanecarboxylic acid ethyl ester hydrochloride 42 (4.92 g, 31.7 mmol) and HATU (12.6 g, 33.2 mmol) were added to 41 (8.14 g, 30.2 mmol). The mixture was cooled in an ice bath under argon, and then DMF (100 mL) and DIPEA (12.5 mL, 11.5 mmol) were successively added. After 30 min at 0°C, the solution was stirred at room temperature for an additional 3 h. Then, the reaction mixture was partitioned between EtOAc and water, washed successively with 0.5 N HCl (20 mL) and saturated NaCl (2 x 20 mL), and dried (Na2SO4). Purification by flash chromatography (AcOEt/CH2Cl2/Petroleum ether, 1:1:1) afforded 7.41 g (60%) of the target product 43 as a colorless oil: m/z = 407 (M+H)+.

DIAD (1.02 mL, 5.17 mmol) was added at -15°C under nitrogen atmosphere to a solution of 43 (1.5 g, 3.69 mmol), quinoline 36 (1.39 g, 4.43 mmol) and triphenyl- phosphine (1.26 g, 4.80 mmol) in dry THF (40 mL). After 4.5 h, at -15°C, the reaction mixture was partitioned between ice-cold water and AcOEt, dried (Na2SO4) and evaporated. The crude material was purified by flash column chromatography (gradient of petroleum AcOEt/CH2Cl2, 1 :9 to 2:8) to give 1.45 g (56 %) of the target product 44: m/z = 703 (M+H)+.

A solution of 44 (1.07 g, 1.524 mmol) and Hoveyda-Grubbs 1st generation catalyst (33 mg, 0.03 eq) in dried and degassed 1,2-dichloroethane (900 mL) was heated at 75°C under nitrogen for 12 h. Then, the solvent was evaporated and the residue purified by silica gel chromatography (25% EtOAc in CH2Cl2). 620 mg (60%) of pure macrocycle 45 were obtained, m/z = 674 (M+H)+. 1H NMR (CDCl3): 1.18-1.39 (m, 12H), 1.59 (m, IH), 1.70-2.08 (m, 5H), 2.28 (m, IH), 2.38 (m, IH), 2.62 (m, 2H), 2.68 (s, 3H), 2.83 (m, IH), 3.06 (s, 3H), 3.19 (sept, J= 6.7 Hz, IH), 3.36 (m, IH), 3.83 (m, IH), 3.97 (s, 3H), 4.09 (m, 2H), 4.65 (td, J= 4 Hz, 14 Hz, IH), 5.19 (dd, J= 4 Hz, 10 Hz, IH), 5.31 (m, IH), 5.65 (td, J= 4 Hz, 8 Hz, IH), 7.00 (s, IH), 7.18 (s, IH), 7.46 (d, J= 9 Hz, IH), 7.48 (s, IH), 8.03 (d, J= 9 Hz, IH).
Step F

A solution of lithium hydroxide (1.65 g, 38.53 mmol) in water (15 mL) was added to a stirred solution of ester 45 (620 mg, 0.920 mmol) in THF (30 mL) and MeOH (20 mL). After 16 h at room temperature, the reaction mixture was quenched with NH4Cl sat., concentrated under reduced pressure, acidified to pH 3 with HCl IN and extracted with CH2Cl2, dried (MgSO4) and evaporated to give 560 mg (88%) of carboxylic acid 46. m/z = 647 (M+H)+. 1H NMR (CDCl3): 1.11-1.40 (m, 8H), 1.42-1.57 (m, 2H), 1.74 (m, 2H), 1.88-2.00 (m, 2H), 2.13 (m, IH), 2.28 (m, IH), 2.40 (m, IH), 2.59 (m, 2H), 2.67 (s, 3H), 2.81 (m, IH), 2.97 (s, 3H), 3.19 (m, IH), 3.31 (m, IH), 3.71 (m, IH), 3.96 (s, 3H), 4.56 (dt, J= 4 Hz, 12 Hz, IH), 5.23 (m, 2H), 5.66 (m, IH), 7.01 (s, IH), 7.10 (s, IH), 7.22 (d, J= 10 Hz, IH), 7.45 (s, IH), 8.00 (d, J= 10 Hz, IH).
Example 5: Preparation of JV-ri7-r2-(4-isopropylthiazole-2-yl)-7-methoxy-8- methylquinolin-4- yloxyl – 13 -methyl-2, 14-dioxo-3 , 13 -diazatricyclol“ 13.3.0.04‘6loctadec- 7-ene-4-carbonyll (cvclopropyPsulfonamide (47) SIMEPREVIR

A solution of 17-[2-(4-isopropylthiazole-2-yl)-7-methoxy-8-methylquinolin-4-yloxy]- 13-methyl-2, 14-dioxo-3, 13-diazatricyclo[l 3.3.0.04,6]octadec-7-ene-4-carboxylic acid 46 (560mg, 0.867 mmol) prepared according to Example 4, and carbonyldiimidazole (308 mg, 1.90 mmol) in dry THF (10 mL) was stirred at reflux under nitrogen for 2h. The reaction mixture was cooled to room temperature and cyclopropylsulfonamide (400 mg, 3.301 mmol) and DBU (286 mg, 1.881 mmol) were added. This solution was heated at 50°C for 15 h. Then, the reaction mixture was cooled down at room temperature and concentrated under reduced pressure. The residue was partitioned between CH2CI2 and HCl 1 N, the organic layer was washed with brine, dried (MgSO4) and evaporated. Purification by flash chromatography (gradient of EtOAc (0 to 25%) in CH2CI2) afforded 314 mg of an off-white solid which was further washed with water, then isopropylether, and dried in the vacuum oven to deliver 282 mg (40%) of the pure title product 47 SIMEPREVIR as a white powder: m/z = 750 (M+H)+.
1H NMR (CDCl3): 0.99-1.52 (m, 14H), 1.64-2.05 (m, 4H), 2.77 (m, IH), 2.41 (m, 2H), 2.59 (m, 2H), 2.69 (s, 3H), 2.92 (m, 2H), 3.04 (s, 3H), 3.19 (m, IH), 3.40 (m, 2H), 3.98 (s, 3H), 4.60 (t, J= 13 Hz, IH), 5.04 (t, J= 11 Hz, IH), 5.37 (m, IH), 5.66 (m, IH), 6.21 (s, IH), 7.02 (s, IH), 7.22 (d, J= 10 Hz, IH), 7.45 (s, IH), 7.99 (d, J= 10 Hz, IH), 10.82 (broad s, IH).
…………………..
REFERENCES
- “Medivir Announces That Simeprevir (TMC435) Data Will Be Presented at the Upcoming AASLD Meeting”. Yahoo News. October 1, 2012. Retrieved November 6, 2012.
- Lin, TI; Lenz, O; Fanning, G; Verbinnen, T; Delouvroy, F; Scholliers, A; Vermeiren, K; Rosenquist, A et al. (2009). “In vitro activity and preclinical profile of TMC435350, a potent hepatitis C virus protease inhibitor”. Antimicrobial agents and chemotherapy 53 (4): 1377–85. doi:10.1128/AAC.01058-08. PMC 2663092. PMID 19171797.
|displayauthors=suggested (help) - “Phase 3 Studies Show Simeprevir plus Interferon/Ribavirin Cures Most Patients in 24 Weeks”. hivandhepatitis.com. December 27, 2012.
- Medivir announces TMC435 in an expanded clinical collaboration. Medivir. 18 April 2012.
- Results from a phase IIa study evaluating Simeprevir and Sofosbuvir in prior null responder Hepatitis C patients have been presented at CROI. 6 March 2013.
- TMC-435350
Drugs Fut 2009, 34(7): 545 - Structure-activity relationship study on a novel series of cyclopentane-containing macrocyclic inhibitors of the hepatitis C virus NS3/4A protease leading to the discovery of TMC435350
Bioorg Med Chem Lett 2008, 18(17): 4853 - Synthesis of enantiomerically pure trans-3,4-substituted cyclopentanols by enzymatic resolution
Acta Chem Scand (1989) 1992, 46: 1127
PATENTS
- WO 2008092954
- WO 2007014926
- WO 2008092955
- WO 2000009543
- CN 102531932
- WO 2013061285
- WO 2011113859
- WO 2013041655
| WO2010097229A2 * | 26 Feb 2010 | 2 Sep 2010 | Ortho-Mcneil-Janssen Pharmaceuticals Inc | Amorphous salt of a macrocyclic inhibitor of hcv |
| WO2013037705A2 * | 7 Sep 2012 | 21 Mar 2013 | Fovea Pharmaceuticals | Aniline derivatives,their preparation and their therapeutic application |
| WO2005073195A2 * | 28 Jan 2005 | 11 Aug 2005 | Per-Ola Johansson | Hcv ns-3 serine protease inhibitors |
| WO2007014926A1 * | 28 Jul 2006 | 8 Feb 2007 | Tibotec Pharm Ltd | Macrocyclic inhibitors of hepatitis c virus |

The compound ritonavir, and pharmaceutically acceptable salts thereof, and methods for its preparation are described in WO94/14436. For preferred dosage forms of ritonavir, see US6,037, 157, and the documents cited therein: US5,484, 801, US08/402,690, and WO95/07696 and WO95/09614. Ritonavir has the following formula:

Otsuka and Lundbeck’s Once-Monthly Abilify Maintena(R) (Aripiprazole) Now Approved in Europe
Otsuka and Lundbeck’s Once-Monthly Abilify Maintena(R) (Aripiprazole) Now Approved in Europe for Maintenance Treatment of Schizophrenia in Adult Patients Stabilized with Oral Aripiprazole
do not forget to see my old blog post, contains synthesis
Preventing relapse is critical in the treatment of schizophrenia. Pivotal
      studies, that supported the EU submission, demonstrate that Abilify
      Maintena can reduce the risk of relapse relative to placebo and is
      non-inferior to oral aripiprazole in patients with schizophrenia1,2

   -- The approval of Abilify Maintena now provides patients with schizophrenia
      access to a once-monthly formulation with established efficacy together
      with a tolerability profile that is comparable to that of oral ABILIFY(R)
      (aripiprazole)2

   -- At the end of the randomized, double-blind treatment phase in one of the
      pivotal trials, 93 percent of patients reported being extremely, very or
      somewhat satisfied with Abilify Maintena treatment3

   -- Abilify Maintena is the only dopamine D2 partial agonist in a
      once-monthly, injectable form to receive marketing authorization in
      Europe for maintenance treatment of schizophrenia

   -- Abilify Maintena will be the first commercialized product in Europe from
      the global alliance between Otsuka and Lundbeck which is focused on
      developing Central Nervous System (CNS) therapies worldwide
TOKYO & COPENHAGEN, Denmark--(BUSINESS WIRE)--November 21, 2013--
Otsuka Pharmaceutical Co., Ltd. (Otsuka) and H. Lundbeck A/S (Lundbeck) today announced marketing authorization approval from the European Commission for Abilify Maintena (aripiprazole), an intramuscular (IM) once-monthly injectable formulation for maintenance treatment of schizophrenia in adult patients stabilized with oral aripiprazole.
Abilify Maintena reduces the risk of relapse relative to placebo over the long-term and provides effective treatment of schizophrenia.(1,2) It has a tolerability profile similar to oral aripiprazole(1) , and demonstrated statistically significant benefits on patients’ personal and social functioning as compared to placebo.(1,4) 93 percent of patients treated with Abilify Maintena were extremely, very or somewhat satisfied with their treatment at the end of the double-blind treatment phase.(4)
“We strongly believe the schizophrenia community will welcome the availability of Abilify Maintena to help improve outcomes for patients living with schizophrenia. As a company, our focus is to develop treatments to help protect against relapse and preserve brain function,” said Ole Vahlgren, CEO & President, Otsuka Europe. “We must partner with health care professionals and caregivers to help patients get the best treatments that focus on reducing the risk of relapse.”
“Studies have shown that the early use of long-acting injectables can prevent a person with schizophrenia from experiencing a relapse,”(5) said Ole Chrintz, SVP International Markets & Europe, Lundbeck. “Efficacy is important, but a treatment for a chronic condition such as schizophrenia also needs to be well tolerated so patients will stay on it over the long-term. We believe Abilify Maintena meets this need.”(1,2,3)
About the Studies
The efficacy of Abilify Maintena was demonstrated in two double-blind, Phase III, randomized trials. The safety profile for Abilify Maintena was demonstrated to be similar to that of oral ABILIFY. The most frequently observed adverse drug reactions (ADRs) reported in >=5 percent of patients in two double-blind controlled clinical trials of Abilify Maintena were weight increases (9.0 percent), akathisia (7.9 percent), insomnia (5.8 percent), and injection site pain (5.1 percent).(1,2)
About Abilify Maintena
Abilify Maintena is the only dopamine D2 partial agonist in once-monthly, injectable form to receive marketing authorization for maintenance treatment in schizophrenia. Physicians now have an alternative treatment option, with a tolerability profile comparable to the well-established oral ABILIFY, to address the on-going need to reduce the risk of relapse in patients with schizophrenia.
The European label states that Abilify Maintena is a powder and solvent for prolonged-release suspension for intra-muscular (IM) injection. It is a once-monthly formulation of aripiprazole in a sterile lyophilized powder that is reconstituted with sterile water. Abilify Maintena is indicated for maintenance treatment of schizophrenia in adult patients stabilized with oral aripiprazole.
After the first injection, treatment with 10 mg to 20 mg oral aripiprazole should be continued for 14 consecutive days to maintain therapeutic aripiprazole concentrations during initiation of therapy.
About Schizophrenia and Disease Relapse
Schizophrenia is a disease characterized by a distortion in the process of thinking and of emotional responsiveness. It most commonly manifests as hallucinations, paranoid or bizarre delusions, or disorganized speech and thinking, and is accompanied by significant social or occupational dysfunction. Onset of symptoms typically occurs in young adulthood, and the condition is chronic, often requiring lifelong treatment to mitigate symptoms.
Relapse of schizophrenia refers to an exacerbation or acute psychotic break that is characterised primarily by the emergence of positive symptoms such as hallucinations, delusions and disordered thinking.(6)
Relapse can occur when a patient no longer responds to antipsychotic medication, does not take the medication as prescribed, or stops taking their medication altogether. There are many reasons patients stop taking their medication, including poor insight about their illness, side effects from current treatments, complicated medication regimen or lack of support from family.(7) Abilify Maintena is able to significantly reduce the risk of relapse in patients with schizophrenia.(1)
It has been estimated that schizophrenia affects approximately one percent of the adult population in the U.S. and Europe, and approximately 24 million people worldwide.(8,9) In Europe there are approximately 4.4 million adults with schizophrenia,(10) prevalent equally in both genders.(11,12) While there is no cure for the disease, symptoms and risk of relapse can be managed in most patients with appropriate antipsychotic treatment. However, when the disease is not managed, patients are at increased risk of disease relapse, which can cause the re-emergence or worsening of psychotic symptoms.(13)
Schizophrenia places a significant burden on society. It is regarded among the most financially costly illnesses and is according to the World Health Organization (WHO), the 8(th) leading cause of DALYs (lost healthy years) worldwide among patients between the age of 15-44.(11) With 50 percent of patients not receiving appropriate care and 80 percent of patients relapsing within the first 5 years,(14) there is a significant unmet need to be addressed in schizophrenia.
About the Lundbeck and Otsuka Global Alliance
Lundbeck and Otsuka established a global alliance in November 2011 to bring to bear their considerable experience and resources in the CNS area to introduce next-generation treatments for conditions such as schizophrenia, depression, Alzheimer’s disease and alcohol dependency.
About Otsuka Pharmaceutical Co., Ltd.
Otsuka Pharmaceutical Co., Ltd. is a global healthcare company with the corporate philosophy: ‘Otsuka-people creating new products for better health worldwide.’ Otsuka researches, develops, manufactures and markets innovative and original products, with a focus on pharmaceutical products for the treatment of diseases and nutraceutical products for the maintenance of everyday health.
In pharmaceuticals, Otsuka is a leading firm in the challenging area of mental health and also has research programs for several under-addressed diseases including tuberculosis, a significant global public health issue. These commitments illustrate more powerfully than words how Otsuka is a “big venture” company at heart, applying a youthful spirit of creativity in everything it does.
Otsuka is a wholly owned subsidiary of Otsuka Holdings Co., Ltd., the holding company for the Otsuka Group. The chairman Akihiko Otsuka is the third generation of Otsuka family members to lead the business, whose origins date from 1921. The Otsuka Group employs approximately 42,000 people globally and its products are available in more than 80 countries worldwide. Consolidated sales were approximately EUR10 billion or USD 13 billion for fiscal year 2012 (4/1/2012-3/31/2013). Otsuka Pharmaceutical welcomes you to visit its global website at https://www.otsuka.co.jp/en/
About H. Lundbeck A/S
Lundbeck is a global pharmaceutical company highly committed to improving the quality of life of people living with brain diseases. For this purpose, Lundbeck is engaged in the entire value chain throughout research, development, production, marketing and sales of pharmaceuticals across the world. The company’s products are targeted at disorders such as depression and anxiety, psychotic disorders, epilepsy, Huntington’s, Alzheimer’s and Parkinson’s diseases. Lundbeck’s pipeline consists of several mid- to late-stage development programmes.
Lundbeck employs more than 5,800 people worldwide, 2,000 of whom are based in Denmark. We have research in 57 countries and our products are registered in more than 100 countries. We have research centers in Denmark, China and the United States and production facilities in Italy, France, Mexico, China and Denmark. Lundbeck generated revenue of approximately DKK 15 billion in 2012. For additional information, we encourage you to visit our corporate site http://www.lundbeck.com.
REFERENCES:
1.   Kane, JM et al. Aripiprazole intramuscular depot as maintenance treatment in
     patients with schizophrenia: a 52-week, multicenter, randomized,
     double-blind, placebo-controlled study. J Clin Psychiatry 2012;73(5):617-62
2.   Fleischhacker WW, Sanchez R, Perry PP, et al. Aripiprazole once-monthly for
     the treatment of schizophrenia: a double-blind, randomized, non-inferiority
     study vs. oral aripiprazole. Annual Meeting of the American Psychiatric
     Association, 18--22 May, 2013 (poster).
3.   Sanchez R, et al. Patient-reported Outcomes with
     Aripiprazole-Intramuscular-Depot for Long-term Maintenance Treatment in
     Schiziphrenia. NR6-42 2012 (poster)
4.   Carson WH, Perry P, Sanchez R, et al. Effects of a Once-Monthly Formulation
     of Aripiprazole on Secondary Efficacy Outcomes in Maintenance Treatment of
     Schizophrenia. Institute on Psychiatric Services meeting, October 4--7, 2012
     (Poster)
5.   Long-acting injectable antipsychotics in the treatment of schizophrenia:
     their role in relapse prevention. Agid O, et al. Expert Opin. Pharmacother.
     (2010) 11(14):2301-2317
6.   Ayuso-Gutierrez JL, del Rio Vega JM. Factors influencing relapse in the
     long-term course of schizophrenia. Schizophr Res. 1997; 28(2-3): 199-206
7.   Baloush-Kleinman V, et al. Adherence to antipsychotic drug treatment in
     early-episode schizophrenia: a six-month naturalistic follow-up study.
     Schizophr Res. 2011;130(1-3):176-81.
8.   National Institute of Mental Health (NIMH). Health Topics: Statistics.
     Available at http://www.nimh.nih.gov/statistics/1SCHIZ.shtml. Accessed
     October 22, 2013
9.   World Health Organization (WHO). Schizophrenia Fact Sheet. 2010. Available
     at: http://www.who.int/mental_health/management/schizophrenia/en/. Accessed
     October 22, 2013.
10.  World Health Organization (WHO) The global burden of disease:2004 update
     (2008)
11.  National Institute of Mental Health (NIMH). The Numbers Count: Mental
     Disorders in America. 2010. Available at
     http://www.nimh.nih.gov/health/publications/the-numbers-count-mental-disorde
     rs-in-America/index.shtml#Schizophrenia. Accessed October 22, 2013
12.  Regier DA, et al. The de facto US mental and addictive disorders service
     system. Epidemiologic catchment area prospective 1-year prevalence rates of
     disorders and services. Arch Gen Psychiatry. 1993;50(2):85-94.
13.  American Psychiatric Association. Practice guideline for the treatment of
     patients with schizophrenia. Second edition. 2004. Available at
     http://psychiatryonline.org/content.aspx?bookid=28§ionid=1665359.
     Accessed October 28, 2013
14.  Robinson D, et al. Predictors of relapse following response from a first
     episode of schizophrenia or schizoaffective disorder. Arch Gen Psychiatry.
     1999;56(3):241-247


read my earlier blog post
https://newdrugapprovals.wordpress.com/2013/02/07/the-european-medicines-agency-ema-approves-otsukas-aripiprazole-abilify-for-the-treatment-of-moderate-to-severe-manic-episodes-in-bipolar-i-disorder-in-adolescents/
Bayer, Onyx win early FDA OK for Nexavar (sorafenib) in thyroid cancer
The U.S. Food and Drug Administration said on Friday it has expanded the approved use of the cancer drug Nexavar to include late-stage differentiated thyroid cancer.
Differentiated thyroid cancer is the most common type of thyroid cancer, the FDA said. The National Cancer Institute estimates that 60,220 people in the United States will be diagnosed with it and 1,850 will die from the disease in 2013.
The drug, made by Germany’s Bayer AG and Onyx Pharmaceuticals, is already approved to treat advanced kidney cancer and liver cancer that cannot be surgically removed. Onyx was acquired by Amgen Inc earlier this year.
READ ABOUT SORAFENIB IN MY EARLIER BLOGPOST
https://newdrugapprovals.wordpress.com/2013/07/16/nexavar-sorafenib/
SCRIP Awards 2013 -Best Company in an Emerging Market – Dr Reddy’s Laboratories – India, Novartis’s Bexsero, Best New Drug

The SCRIP Awards 2013 celebrated achievements in the global biopharma industry last night at the Lancaster, London.
Hosted by Justin Webb, the evening was a fantastic mix of dining, entertainment and awards.
Among the winners were:
- Novartis’s Bexsero, Best New Drug
- Genmab, Biotech Company of the Year
- Regeneron Pharmaceuticals and Sanofi’s Phase IIa study dupilumab in asthma, Clinical Advance of the Year
You can view the full roll of honour by clicking on the button below.
It was a great night and we would like to thank all those who entered and attended this year’s awards.
Finally congratulations to our winners and a huge thanks to our sponsors for helping us make it such a fantastic success.
Don’t forget to check our website in the next couple of days for all the pictures from the night.
2013 Winners
Best Company in an Emerging Market – Sponsored by Clinigen Group
- Dr Reddy’s Laboratories – India
Best Technological Development in Clinical Trials
- Quintiles’s Infosario Safety
Best Partnership Alliance
- AstraZeneca with Bristol-Myers Squibb and Amylin in diabetes
Financing Deal of the Year
- Mesoblast’s equity financing of Aus$170m
Best Advance in an Emerging Market
- Novartis’s Jian Kang Kuai Che Healthcare Project in China
Clinical Advance of the Year – Sponsored by Quintiles
- Regeneron Pharmaceuticals and Sanofi’s Phase IIa study dupilumab in in asthma
Licensing Deal of the Year – Sponsored by Hume Brophy
- AstraZeneca and Horizon Discovery for the development and commercialization of the HD-001 kinase target program for multiple cancer types
Executive of the Year
- Roch Doliveux, chairman and chief executive officer of UCB
Biotech Company of the Year
- Genmab
Best Contract Research Organization
- Quintiles
Management Team of the Year
- Regeneron Pharmaceuticals’ CEO Leonard S Schleifer and CSO George D Yancopoulos
Best New Drug – Sponsored by INC Research
- Novartis’ Bexsero (meningococcal group B vaccine)
Pharma Company of the Year – Sponsored by ICON
- Astellas
Lifetime Achievement Award
- Prof Dr Désiré Collen
…….read about bexero at

DR ANTHONY MELVIN CRASTO Ph.D


