PROUD Indian WORLD RECORD VIEWS holder on THIS BLOG, ………live, by DR ANTHONY MELVIN CRASTO, Worldpeaceambassador, Worlddrugtracker, Helping millions, 100 million hits on google, pushing boundaries,2.5 lakh plus connections worldwide, 45 lakh plus VIEWS on this blog in 227 countries, 7 CONTINENTS ……A 90 % paralysed man in action for you, I am suffering from transverse mylitis and bound to a wheel chair, [THIS BLOG HOLDS WORLD RECORD VIEWS ] A PROUD INDIAN
DR ANTHONY MELVIN CRASTO, Born in Mumbai in 1964 and graduated from Mumbai University, Completed his Ph.D from ICT, 1991,Matunga, Mumbai, India, in Organic Chemistry, The thesis topic was Synthesis of Novel Pyrethroid Analogues, Currently he is working with AFRICURE PHARMA, ROW2TECH, NIPER-G, Department of Pharmaceuticals, Ministry of Chemicals and Fertilizers, Govt. of India as ADVISOR, earlier assignment was
with GLENMARK LIFE SCIENCES LTD, as CONSUlTANT, Retired from GLENMARK in Jan2022 Research Centre as Principal Scientist, Process Research (bulk actives) at Mahape, Navi Mumbai, India. Total Industry exp 32 plus yrs, Prior to joining Glenmark, he has worked with major multinationals like Hoechst Marion Roussel, now Sanofi, Searle India Ltd, now RPG lifesciences, etc. He has worked with notable scientists like Dr K Nagarajan, Dr Ralph Stapel, Prof S Seshadri, etc, He did custom synthesis for major multinationals in his career like BASF, Novartis, Sanofi, etc., He has worked in Discovery, Natural products, Bulk drugs, Generics, Intermediates, Fine chemicals, Neutraceuticals, GMP, Scaleups, etc, he is now helping millions, has 9 million plus hits on Google on all Organic chemistry websites. His friends call him Open superstar worlddrugtracker. His New Drug Approvals, Green Chemistry International, All about drugs, Eurekamoments, Organic spectroscopy international,
etc in organic chemistry are some most read blogs He has hands on experience in initiation and developing novel routes for drug molecules
and implementation them on commercial scale over a 32 PLUS year tenure till date Feb 2023, Around 35 plus products in his career. He has good knowledge of IPM, GMP, Regulatory aspects, he has several International patents published worldwide . He has good proficiency in Technology transfer, Spectroscopy, Stereochemistry, Synthesis, Polymorphism etc., He suffered a paralytic stroke/ Acute Transverse mylitis in Dec 2007 and is 90 %Paralysed, He is bound to a wheelchair, this seems to have injected feul in him to help chemists all around the world, he is more active than before and is pushing boundaries, He has 100 million plus hits on Google, 2.5 lakh plus connections on all networking sites, 100 Lakh plus views on dozen plus blogs, 227 countries, 7 continents, He makes himself available to all, contact him on +91 9323115463, email amcrasto@gmail.com, Twitter, @amcrasto , He lives and will die for his family, 90% paralysis cannot kill his soul., Notably he has 38 lakh plus views on New Drug Approvals Blog in 227 countries......https://newdrugapprovals.wordpress.com/ , He appreciates the help he gets from one and all, Friends, Family, Glenmark, Readers, Wellwishers, Doctors, Drug authorities, His Contacts, Physiotherapist, etc
He has total of 32 International and Indian awards
Bulevirtide-gmod, sold under the brand name Hepcludex, is the first and only FDA-approved medication for treating chronic hepatitis delta virus (HDV) infection in adults. Developed by Gilead Sciences, it received accelerated approval from the U.S. Food and Drug Administration (FDA) on May 22, 2026, filling a critical gap for patients with this severe viral liver disease.
Indication and Clinical Use
Target Patient Profile: Approved for adults with chronic HDV who have compensated cirrhosis or no cirrhosis.
The Clinical Need: HDV only occurs as a co-infection in individuals who already have Hepatitis B (HBV). It is considered the most aggressive form of viral hepatitis, often accelerating liver scarring (fibrosis), liver failure, and liver cancer.
Basis of Approval: The FDA granted accelerated approval based on Phase 3 MYR301 study data, which demonstrated a significant reduction in viral HDV RNA and the normalization of alanine aminotransferase (ALT) liver enzymes.
Mechanism of Action
Bulevirtide-gmod is a first-in-class entry inhibitor. It works by binding to and blocking the sodium taurocholate co-transporting polypeptide (NTCP) receptor on liver cells. Because HDV and HBV rely on this specific receptor to enter hepatocytes, the drug successfully disrupts the viral life cycle and prevents the virus from spreading to healthy liver cells.
Dosage and Administration
Form: Supplied as a lyophilized powder for injection.
Dose: The recommended dose is 8.5 mg once daily.
Administration: Delivered via subcutaneous injection (under the skin).
Safety and Side Effects
Boxed Warning: The drug carries a prominent warning regarding the risk of severe acute exacerbations of hepatitis D and B if treatment is discontinued. Stopping the medication can cause severe, life-threatening viral flares, requiring close medical monitoring for at least 6 months post-treatment.
Common Side Effects: The most frequent adverse reactions of patients) include:
The most common side effects include raised levels of bile salts in the blood and reactions at the site of injection.[8]
Bulevirtide works by attaching to and blocking a receptor (target) through which the hepatitis delta and hepatitis B viruses enter liver cells.[8] By blocking the entry of the virus into the cells, it limits the ability of HDV to replicate and its effects in the body, reducing symptoms of the disease.[8]
Bulevirtide was approved for medical use in the European Union in July 2020,[8] and in Canada in August 2025.[5]
Medical uses
Bulevirtide is indicated for the treatment of chronic hepatitis delta virus (HDV) infection in plasma (or serum) HDV-RNA positive adult patients with compensated liver disease.[8][10]
The hepatitis B virus uses its surface lipopeptide pre-S1 for docking to mature liver cells via their sodium/bile acid cotransporter (NTCP) and subsequently entering the cells. Myrcludex B is a synthetic N-acylated pre-S1[12][13] that can also dock to NTCP, blocking the virus’s entry mechanism.[14]
Bulevirtide is also effective against hepatitis D because the hepatitis D virus uses the same entry receptor as the hepatitis B virus and is only effective in the presence of a hepatitis B virus infection.[14]
Pre-clinical data in mice suggests that pharmacological inhibition of NTCP-mediated bile salt uptake may also be effective to lower hepatic bile salt accumulation in cholestatic conditions. This reduces hepatocellular damage.[15] An increased ratio of phospholipid to bile salts seen in bile upon NTCP inhibition may further contribute to the protective effect as bile salts are less toxic in presence of phospholipids.[16]
Structural formula
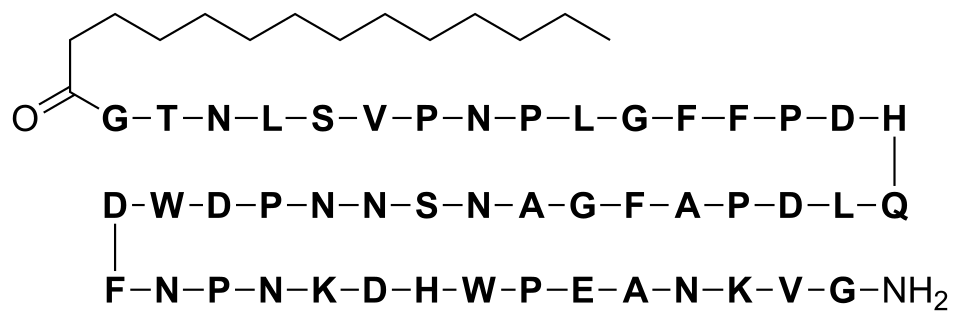
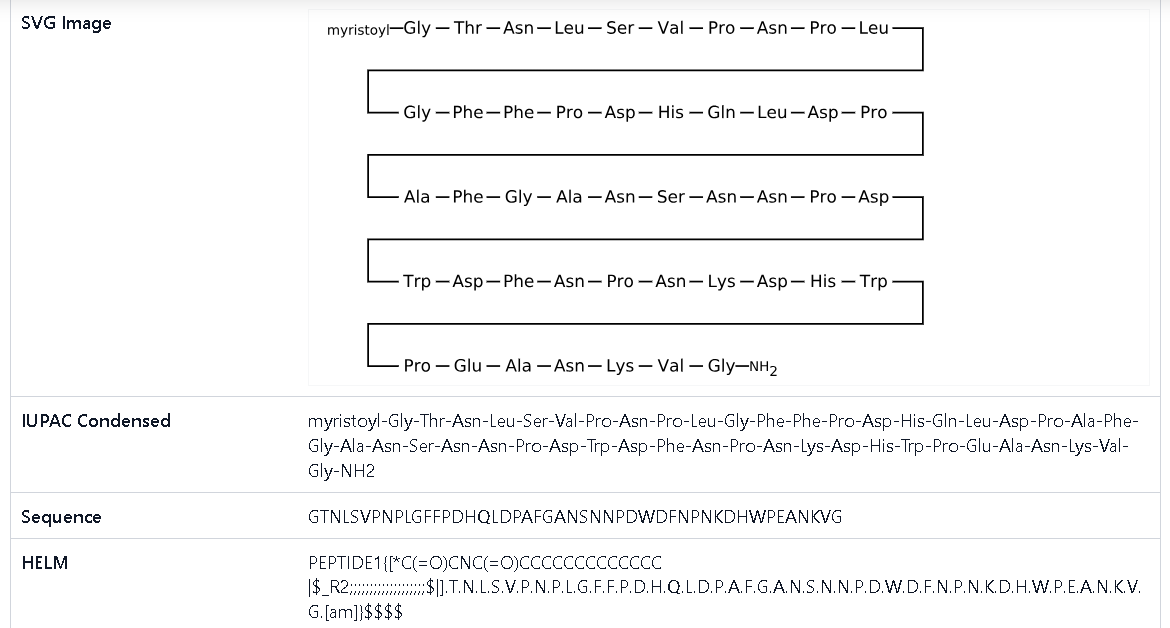
Bulevirtide is a 47-amino acid peptide with the following sequence:[17]
“Hepcludex EPAR”. European Medicines Agency (EMA). 26 May 2020. Retrieved 12 August 2020. Text was copied from this source which is copyright European Medicines Agency. Reproduction is authorized provided the source is acknowledged.
Volz T, Allweiss L, Ben MBarek M, Warlich M, Lohse AW, Pollok JM, et al. (May 2013). “The entry inhibitor Myrcludex-B efficiently blocks intrahepatic virus spreading in humanized mice previously infected with hepatitis B virus”. Journal of Hepatology. 58 (5): 861–867. doi:10.1016/j.jhep.2012.12.008. PMID23246506.
Spreitzer H (14 September 2015). “Neue Wirkstoffe – Myrcludex B”. Österreichische Apothekerzeitung (in German) (19/2015): 12.
Na+ -taurocholate cotransporting polypeptide inhibition has hepatoprotective effects in cholestasis in mice. Slijepcevic D, Roscam Abbing RLP, Fuchs CD, Haazen LCM, Beuers U, Trauner M, Oude Elferink RPJ, van de Graaf SFJ. Hepatology. 2018 Sep;68(3):1057-1069. doi: 10.1002/hep.29888
To treat adults with relapsed or refractory mantle cell lymphoma after at least two lines of systemic therapy, including a Bruton’s tyrosine kinase inhibitor
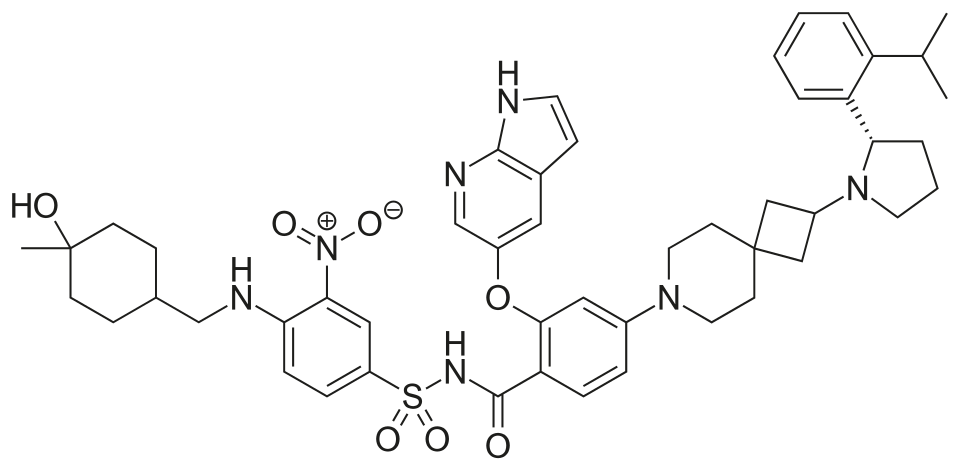
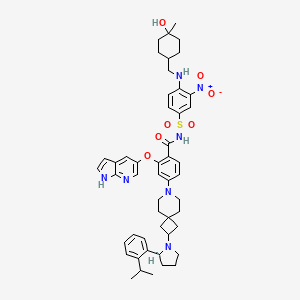
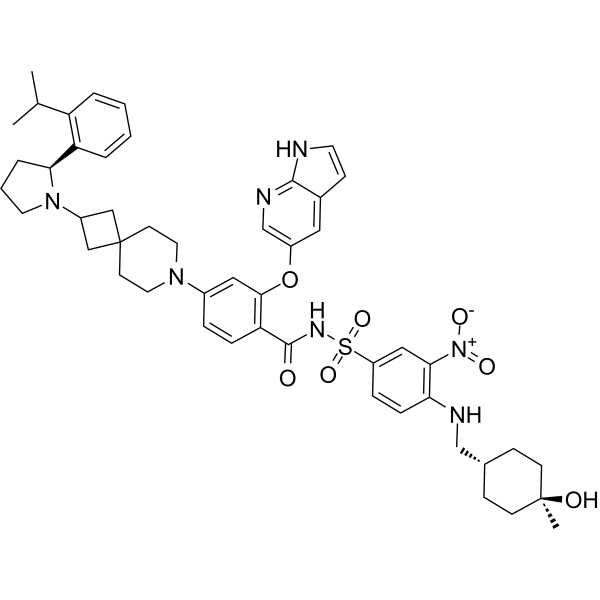
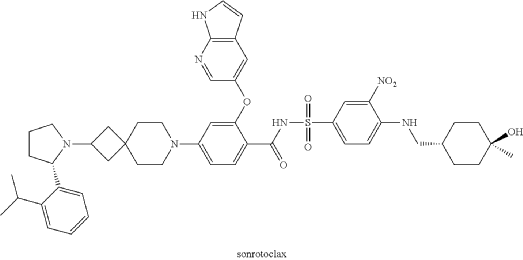
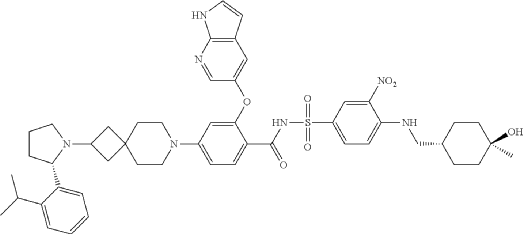
Sonrotoclax is a potent, orally active Bcl2 inhibitor. Sonrotoclax has effective cell killing effect against a variety of lymphoma and leukemia cell lines.
Regulatory Status & Primary Indication
On May 13, 2026, the U.S. Food and Drug Administration (FDA) granted accelerated approval to sonrotoclax for treating adult patients with relapsed or refractory mantle cell lymphoma (MCL). [1]
Eligibility Requirement: Patients must have undergone at least two prior lines of systemic therapy, which must include a Bruton’s tyrosine kinase (BTK) inhibitor.
Clinical Performance: In the supporting Phase 1/2 BGB-11417-201 trial, sonrotoclax demonstrated an overall response rate (ORR) of 52% and a median time to response of 1.9 months
Sonrotoclax is an orally bioavailable inhibitor of the anti-apoptotic protein B-cell lymphoma 2 (Bcl-2), with potential pro-apoptotic and antineoplastic activities. Upon oral administration, sonrotoclax specifically binds to and inhibits the activity of the pro-survival protein Bcl-2. This restores apoptotic processes and inhibits cell proliferation in Bcl-2-overexpressing tumor cells. Bcl-2, a protein that belongs to the Bcl-2 family, is overexpressed in various tumor cell types and plays an important role in the negative regulation of apoptosis. Its tumor expression is associated with increased drug resistance and cancer cell survival.
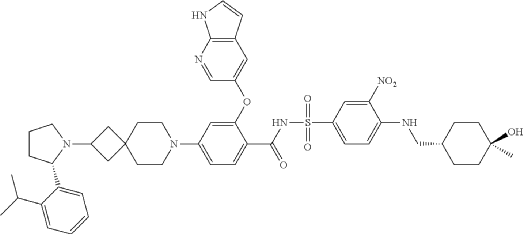
A mixture of (S)-2-((1H-pyrrolo[2,3-b]pyridin-5-yl)oxy)-4-(2-(2-(2-isopropylphenyl)pyrrolidin-1-yl)-7-azaspiro[3.5]nonan-7-yl)benzoic acid (44 g, 78 mmol), 4-((((1r,4r)-4-hydroxy-4-methylcyclohexyl)methyl)amino)-3-nitrobenzenesulfonamide (26.8 g, 78 mmol), TFA (15.7 g, 156 mmol), EDCl (19.4 g, 101 mmol) and DMAP (19 g, 156 mmol) in anhydrous DCM (880 mL) was stirred overnight at room temperature. The reaction was monitored by HPLC. After starting material of (S)-2-((1H-pyrrolo[2,3-b]pyridin-5-yl)oxy)-4-(2-(2-(2-isopropylphenyl)pyrrolidin-1-yl)-7-azaspiro[3.5]nonan-7-yl)benzoic acid was consumed completely, the reaction mixture was heated to ˜35° C. and N 1,N 1-dimethylethane-1,2-diamine (17.2 g, 195 mmol) was added in one portion. The reaction was stirred for another 12 hours. The mixture was washed twice with 10 wt % aq. AcOH solution (300 mL×2) and then washed with saturated aq. NaHCO 3 (300 mL×2). The organic layer was collected and concentrated to about 90 mL. 22 g of silica gel was added and stirred for 2 hours. After filtration, 180 mL EA was added into the filtrate at reflux and further stirred for 5 hours. After the mixture was cooled to room temperature, the precipitate was filtered and then the wet cake was washed twice with EA (180 mL). After drying in vacuum at 80-90° C., the desired compound was obtained (48 g, yield: 69.5%). 1H NMR (DMSO-d 6) δ ppm: 11.65 (s, 1H), 11.11 (br, 1H), 8.58-8.39 (m, 2H), 8.00 (d, J=2.8 Hz, 1H), 7.74 (d, J=8.8 Hz, 1H), 7.57-7.37 (m, 4H), 7.30-7.10 (m, 3H), 7.00 (d, J=9.2 Hz, 1H), 6.65 (d, J=1.2 Hz, 1H), 6.35 (s, 1H), 6.17 (s, 1H), 4.24 (s, 1H), 3.39-3.20 (m, 5H), 3.04-2.88 (m, 4H), 2.23 (s, 1H), 1.94-1.47 (m, 11H), 1.44-1.26 (m, 7H), 1.19 (d, J=8.0 Hz, 3H), 1.14 (d, J=8.0 Hz, 3H), 1.10 (s, 4H). MS (ESI, m/e) [M+1] + 889.9.
Example F43: 2-((1H-pyrrolo[2,3-b]pyridin-5-yl)oxy)-N-((4-((((1r,4r)-4-hydroxy-4-methylcyclohexyl)methyl)amino)-3-nitrophenyl)sulfonyl)-4-(2-((S)-2-(2-isopropylphenyl)pyrrolidin-1-yl)-7-azaspiro[3.5]nonan-7-yl)benzamide
To treat estrogen receptor-positive, human epidermal growth factor receptor 2-negative, ESR1-mutated advanced or metastatic breast cancer with disease progression following at least one line of endocrine therapy
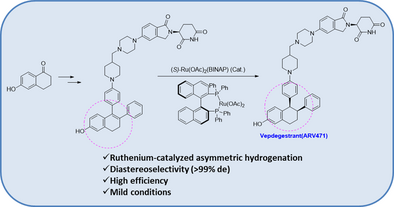
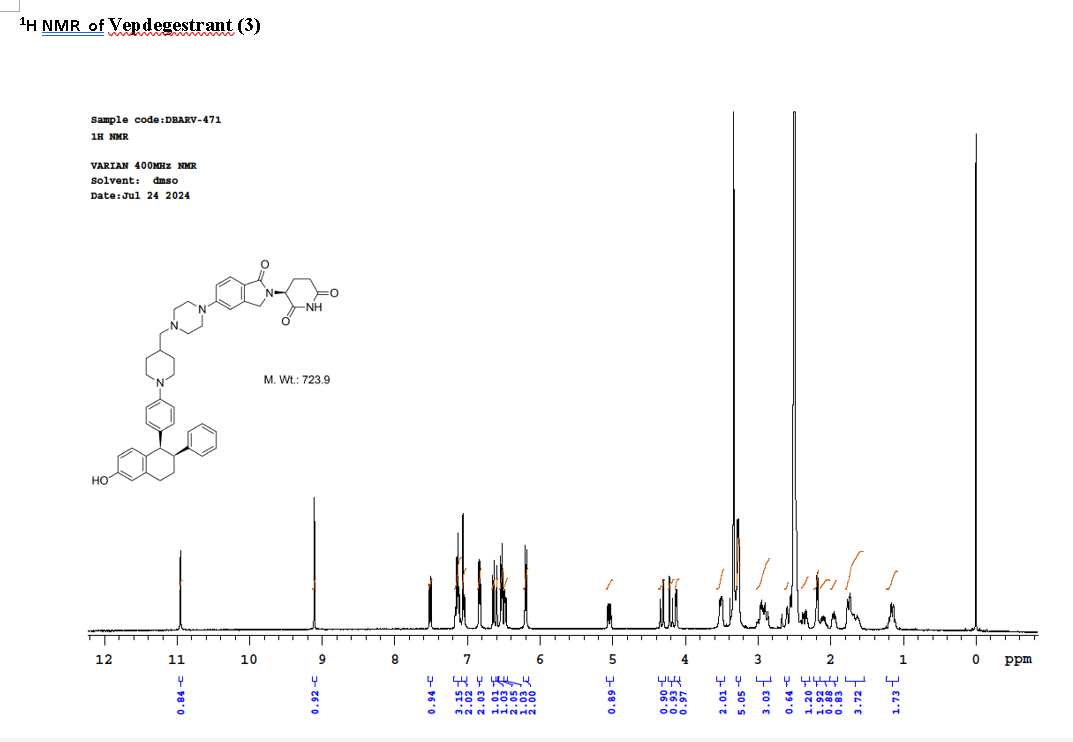
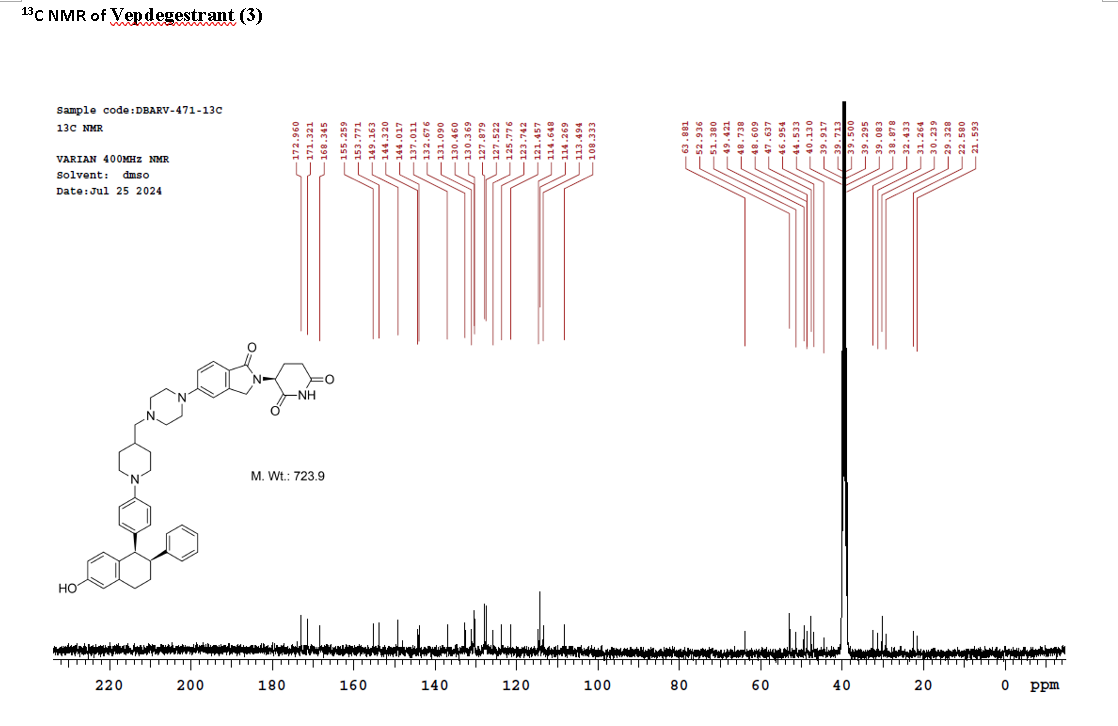
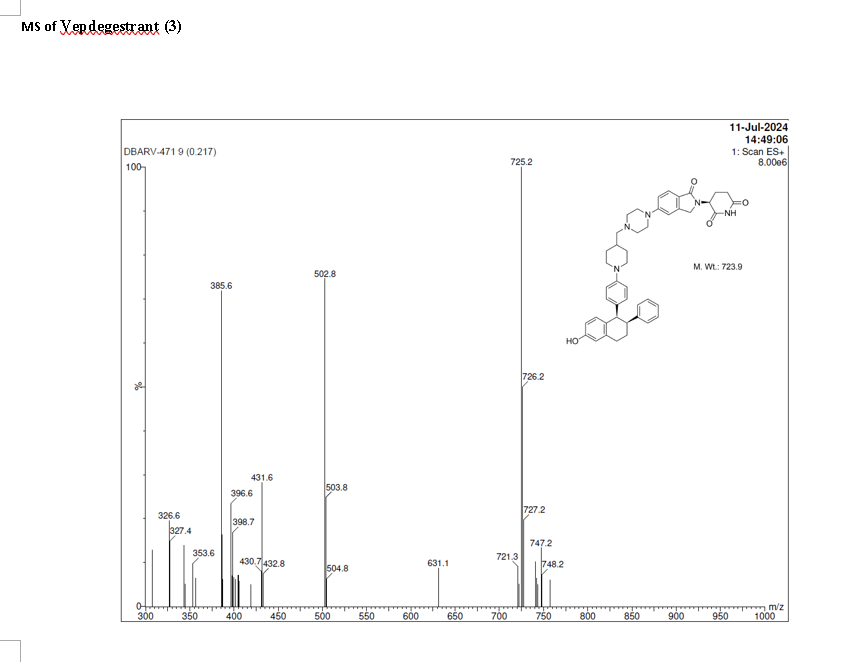
On May 1, 2026, the FDA approved vepdegestrant (Veppanu), a first-in-class oral PROTAC estrogen receptor (ER) degrader developed by Arvinas and Pfizer, for adults with ER-positive, HER2-negative, ESR1-mutated advanced or metastatic breast cancer who have progressed on endocrine therapy. It demonstrated significant progression-free survival (PFS) improvements compared to fulvestrant.
Key Details About Vepdegestrant (Veppanu):
Mechanism of Action: As an oral PROTAC (Proteolysis-Targeting Chimera), vepdegestrant targets the estrogen receptor for degradation, designed to be more effective than traditional endocrine therapies, particularly in ESR1-mutated tumors.
Approved Indication: For treating adults with ER+/HER2-, ESR1-mutated advanced/metastatic breast cancer (detected by Guardant360 CDx) after at least one line of endocrine therapy.
Dosage: The recommended dose is 200 mg taken orally once daily with food.
Clinical Efficacy (VERITAC-2): In trials, vepdegestrant showed a significantly longer PFS compared to intramuscular fulvestrant.
Side Effects & Risks: Common side effects include decreased white blood cell counts, increased liver function tests, muscle/bone pain, fatigue, and nausea. Warnings include embryo-fetal toxicity and QTc interval prolongation (heart rhythm issues).
Companion Diagnostic:Guardant360 CDx was approved alongside the drug to identify patients with ESR1 mutations
Vepdegestrant is designed as a PROTAC that recruits the ubiquitin-proteasome system to target the estrogen receptor for degradation.[4] The compound contains both an E3 ubiquitin ligase-binding moiety and an estrogen receptor-binding domain, intended to bring these proteins into proximity to trigger ubiquitination and subsequent proteasomal degradation of the ER protein.[5] In laboratory studies, vepdegestrant demonstrated ER degradation in ER-positive breast cancer cell lines with reported DC50 values of approximately 1-2 nM.[6]
Vepdegestrant is an orally available hetero-bifunctional molecule and selective estrogen receptor (ER) alpha-targeted protein degrader, using the proteolysis targeting chimera (PROTAC) technology, with potential antineoplastic activity. Vepdegestrant is composed of an ER alpha ligand attached to an E3 ligase recognition moiety. Upon oral administration,vepdegestrant targets and binds to the ER ligand binding domain on ER alpha. E3 ligase is recruited to the ER by the E3 ligase recognition moiety and ER alpha is tagged by ubiquitin. This causes ubiquitination and degradation of ER alpha by the proteasome. This decreases ER alpha protein levels, decreases the expression of ER alpha-target genes and halts ER-mediated signaling. This results in an inhibition of proliferation in ER alpha-overexpressing tumor cells. In addition, the degradation of the ER alpha protein releases the ARV-471 and can bind to additional ER alpha target proteins. ER alpha is overexpressed in a variety of cancers and plays a key role in cancer cell proliferation.
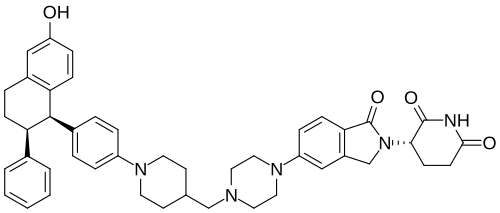
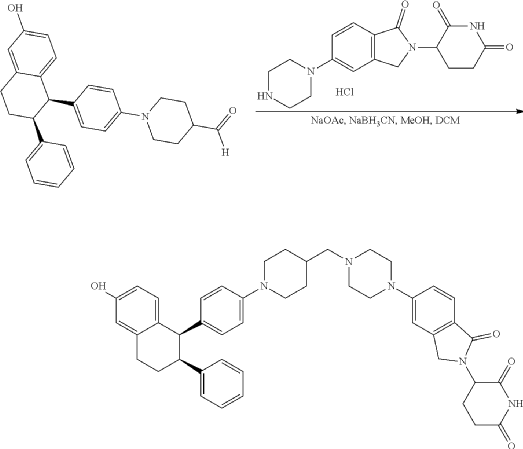
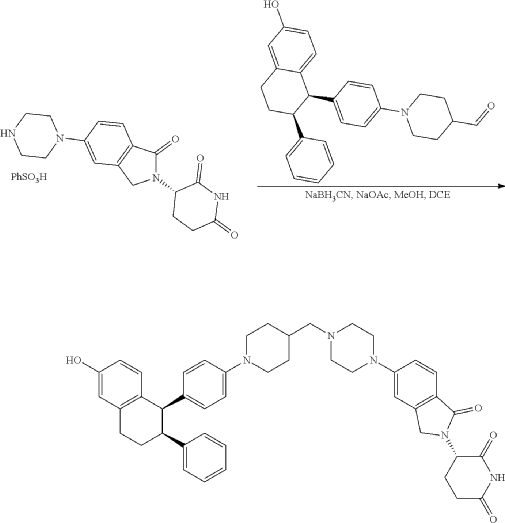
Step 11: Preparation of 3-[5-[4-[[1-[4-[(1R, 2S)-6-hydroxy-2-phenyl-tetralin-1-yl]phenyl]-4-piperidyl]methyl]piperazin-1-yl]-1-oxo-isoindolin-2-yl]piperidine-2,6-dione (Compound (I-b))
To a solution of 3-(1-oxo-5-piperazin-1-yl-isoindolin-2-yl)piperidine-2,6-dione hydrochloride (319 mg, 0.87 mmol, prepared in Step 17 described for Exemplary Compound 62) in methanol (4 mL) and dichloromethane (4 mL) was added sodium acetate (120 mg, 1.46 mmol, 2 eq). The mixture was stirred at 20° C. for 0.5 h, then to the mixture was added 1-[4-[(1R,2S)-6-hydroxy-2-phenyl-tetralin-1-yl]phenyl]piperidine-4-carbaldehyde (300 mg, 0.73 mmol, 1 eq) and sodium cyanoborohydride (137 mg, 2.19 mmol, 3 eq). The mixture was stirred at 20° C. for 12 h. LC-MS showed the starting material was consumed completely and one main peak with desired MW was detected. The reaction mixture was concentrated under reduced pressure. The residue was purified by prep-HPLC (Phenomenex luna C 18 column, 250×50 mm, 10 um; mobile phase: [water (0.05% HCl)-acetonitrile]; B %: acetonitrile 10%-40% in 30 min). The desired compound 3-[5-[4-[[1-[4-[(1R, 2S)-6-hydroxy-2-phenyl-tetralin-1-yl]phenyl]-4-piperidyl]methyl]piperazin-1-yl]-1-oxo-isoindolin-2-yl]piperidine-2,6-dione (288.4 mg, 0.37 mmol, 51% yield) was obtained as a white solid of hydrochloride salt. LC-MS (ESI) m/z: 724.4 [M+1] +; 1H NMR (400 MHz, DMSO-d 6) δ 10.97 (s, 1H), 10.83 (s, 0.9H, HCl), 7.60 (d, J=8.5 Hz, 1H), 7.40 (br s, 2H), 7.22-7.11 (m, 5H), 6.83 (d, J=6.0 Hz, 2H), 6.69-6.63 (m, 2H), 6.58-6.47 (m, 3H), 5.07 (dd, J=5.2, 13.2 Hz, 1H), 4.41-4.30 (m, 2H), 4.28-4.21 (m, 1H), 4.00 (d, J=12.7 Hz, 2H), 3.61 (d, J=11.0 Hz, 2H), 3.54-3.36 (m, 6H), 3.16 (br s, 4H), 3.06-2.84 (m, 3H), 2.76-2.53 (m, 1H), 2.43-2.33 (m, 1H), 2.27 (br s, 1H), 2.16-2.04 (m, 3H), 2.02-1.69 (m, 5H).
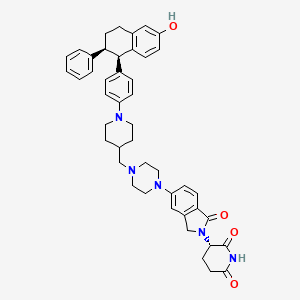
Synthesis of (3S)-3-[5-[4-[[1-[4-[(1R, 2S)-6-hydroxy-2-phenyl-tetralin-1-yl]phenyl]-4-piperidyl]methyl]piperazin-1-yl]-1-oxo-isoindolin-2-yl]piperidine-2,6-dione (Compound (I-c))
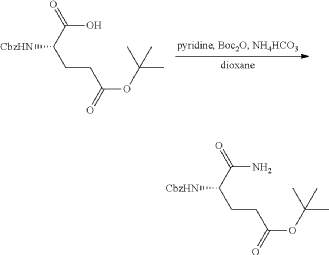
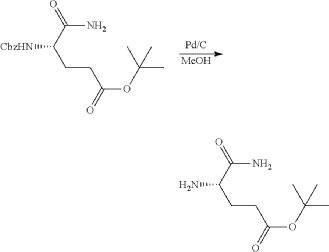
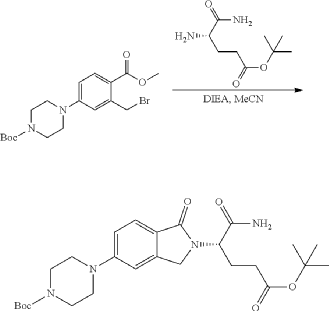
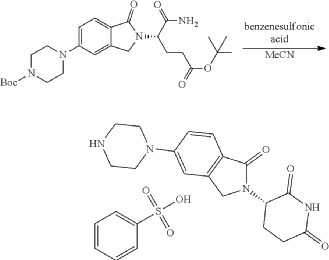
To a mixture of (3 S)-3-(1-oxo-5-piperazin-1-yl-isoindolin-2-yl)piperidine-2,6-dione (1.30 g, 3.47 mmol, 1 eq, benzene sulfonate) in dichloromethane (8 mL) and methanol (32 mL) was added sodium acetate (854 mg, 10.41 mmol, 3 eq) in one portion at 20° C. The mixture was stirred at 20° C. for 10 minutes. Then 1-[4-[(1R, 2S)-6-hydroxy-2-phenyl-tetralin-1-yl]phenyl] piperidine-4-carbaldehyde (1 g, 2.43 mmol, 0.7 eq, prepared as described above in the synthesis of Compound (I-b)) was added. The mixture was stirred at 20° C. for 10 minutes. After that, acetic acid (0.2 mL) and sodium cyanoborohydride (436 mg, 6.94 mmol, 2 eq) was added in one portion. The mixture was stirred at 20° C. for 40 minutes. The mixture was concentrated in vacuum, and 50 mL of tetrahydrofuran and 20 mL of water were added. The mixture was stirred for 20 minutes. Saturated aqueous sodium bicarbonate solution was added to adjust the pH to 8-9. The aqueous phase was extracted with ethyl acetate and tetrahydrofuran (v:v=2:1, 60 mL×3). The combined organic phase was washed with brine (60 mL×1), dried with anhydrous sodium sulfate, filtered and concentrated in vacuum. The residue was purified by preparative reverse phase HPLC (column: Phenomenex luna C18 250×50 mm, 10 micron; mobile phase: [water (0.225% formic acid)-acetonitrile]; B %: 20%-50% in 30 min). The product (3S)-3-[5-[4-[[1-[4-[(1R,2S)-6-hydroxy-2-phenyl-tetralin-1-yl]phenyl]-4-piperidyl]methyl] piperazin-1-yl]-1-oxo-isoindolin-2-yl]piperidine-2,6-dione (964 mg, 1.23 mmol, 35% yield, 98% purity, formate) was obtained as a white solid of formic acid salt after lyophilization. Chiral purity was analyzed by chiral SFC (Chiralcel OJ-3 50×4.6 mm, 3 micron; mobile phase: 50% ethanol (0.05% DEA) in CO 2; flow rate: 3 mL/min, wavelength: 220 nm) and observed t p=2.89 min with de over 95%. [α D=−267.5 (c=0.2 in DMF, 25° C.). LC-MS (ESI) m/z: 724.2 [M+1] +. 1H NMR (400 MHz, DMSO-d 6) δ 10.94 (s, 1H), 8.16 (s, 1H, formate), 7.51 (d, J=8.8 Hz, 1H), 7.21-6.98 (m, 5H), 6.83 (d, J=6.4 Hz, 2H), 6.68-6.57 (m, 2H), 6.56-6.44 (m, 3H), 6.20 (d, J=8.8 Hz, 2H), 5.04 (dd, J=5.2, 13.2 Hz, 1H), 4.32 (d, J=16.8 Hz, 1H), 4.19 (d, J=17.2 Hz, 1H), 4.12 (d, J=4.8 Hz, 1H), 3.51 (br d, J=10.0 Hz, 4H), 3.27 (br s, 8H), 3.03-2.82 (m, 3H), 2.63-2.54 (m, 1H), 2.43-2.28 (m, 2H), 2.19 (d, J=6.8 Hz, 2H), 2.15-2.02 (m, 1H), 2.01-1.89 (m, 1H), 1.83-1.51 (m, 4H), 1.28-1.04 (m, 2H).
Iwata, H.; Naito, Y.; Hattori, M.; Yoshimura, A.; Yonemori, K.; Aizawa, M.; et al. (November 2023). “58P Safety and pharmacokinetics (PK) of vepdegestrant in Japanese patients with estrogen receptor (ER)+/human epidermal growth factor receptor 2 (HER2)- advanced breast cancer: Results from a Japanese phase I study”. Annals of Oncology. 34: S1488–S1489. doi:10.1016/j.annonc.2023.10.193. S2CID265657144.
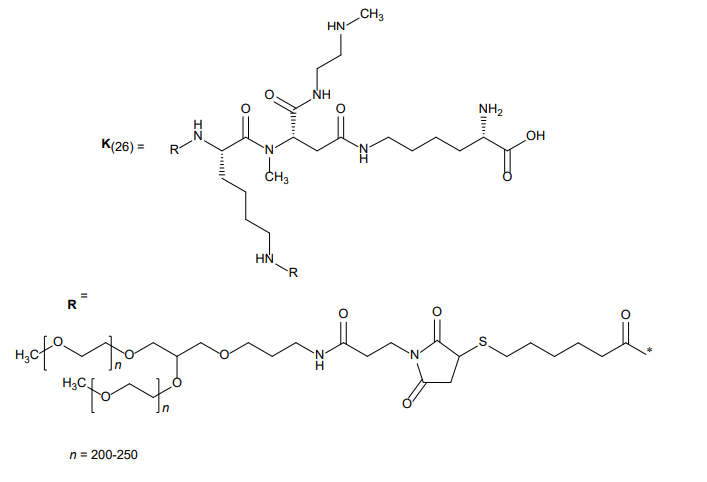
MOLECULAR FORMULA C231H386N64O67S5 + (C2H4O)4n MOLECULAR WEIGHT approx. 45 kDa
The structure of navepegritide (YUVIWEL®) is built using a “prodrug” design. It is not a simple small molecule, but rather a complex conjugate consisting of three distinct components designed to release the active drug slowly over time.
1. The Active Part: C-Type Natriuretic Peptide (CNP)
The core of the molecule is a synthetic 38-amino acid peptide (CNP-38).
Sequence: This peptide mimics the natural human C-type natriuretic peptide, which is essential for bone growth.
Function: Once released, this peptide binds to the natriuretic peptide receptor B (NPR-B) on the surface of chondrocytes (cartilage cells) in the growth plates, stimulating bone formation.
2. The Carrier: Polyethylene Glycol (PEG)
To prevent the body from clearing the small peptide too quickly, it is attached to a large, inert carrier.
Type: It uses a multi-arm, branched 40 kDa Polyethylene Glycol (PEG) molecule.
Purpose: The PEG carrier acts as a shield and a “weight,” making the molecule too large to be filtered out rapidly by the kidneys. This is what allows for once-weekly dosing instead of daily injections.
3. The Linker: TransCon™ Technology
This is the most critical part of the structure. The peptide is attached to the PEG carrier via a cleavable linker.
Mechanism: This linker is designed to break down spontaneously at a predictable rate under physiological conditions (neutral pH and body temperature).
The Result: As the linker slowly breaks, it releases the unmodified, active CNP-38 into the bloodstream. Because the peptide is released in its natural state, it retains its full biological activity.
Summary Table: Structural Components
Component
Description
Role
Peptide
CNP-38 (38 amino acids)
The “payload” that stimulates bone growth.
Linker
pH-sensitive cleavable bond
Controls the slow release of the peptide.
Carrier
40 kDa PEG
Increases the half-life and prevents rapid clearance.
Note: This structure is technically a prodrug because the large PEG-bound version is inactive; only the released CNP-38 peptide performs the therapeutic work.
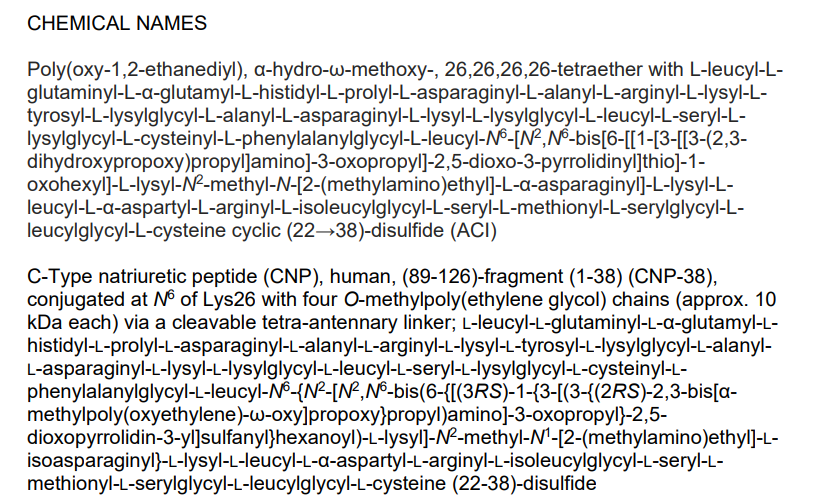
C-Type natriuretic peptide (CNP), human, (89-126)-fragment (1-38) (CNP-38), conjugated at N6 of Lys26 with four O-methylpoly(ethylene glycol) chains (approx. 10 kDa each) via a cleavable tetra-antennary linker; L-leucyl-L-glutaminyl-L-?-glutamyl-L-histid
Poly(oxy-1,2-ethanediyl), ?-hydro-?-methoxy-, 26,26,26,26-tetraether with L-leucyl-L-glutaminyl-L-?-glutamyl-L-histidyl-L-prolyl-L-asparaginyl-L-alanyl-L-arginyl-L-lysyl-L-tyrosyl-L-lysylglycyl-L-alanyl-L-asparaginyl-L-lysyl-L-lysylglycyl-L-leucyl-L-sery
FDA 2026, APPROVALS 2026, 2/27/2026, Yuviwel, Y3BH8M899D, MN-266, TRANSCON CNP, PA (224-233), Influenza, DA-66438, ACP-015, WHO 11981,
To increase linear growth in pediatric patients 2 years and older with achondroplasia with open epiphyses
Navepegritide is a prodrug consisting of a 38-amino acid C-type natriuretic peptide (CNP) moiety conjugated to a multi-arm polyethylene glycol (PEG) carrier via a cleavable linker. This structure allows for the once-weekly dosing approved by the FDA for children with achondroplasia.
Key Details
Purpose: It is designed to increase linear growth by providing continuous exposure to C-type natriuretic peptide (CNP), a protein that helps regulate bone growth.
Mechanism: As a prodrug, it uses Ascendis Pharma’s TransCon technology to release active CNP slowly into the body over a week, maintaining steady levels and avoiding high peaks.
Clinical Benefits: In the pivotal ApproaCH trial, patients treated with navepegritide showed a significant improvement in annualized growth velocity (AGV) compared to those on a placebo. It also showed potential improvements in body proportionality and lower-limb alignment.
Administration: It is administered via a once-weekly subcutaneous injection, offering a less frequent alternative to daily treatments like vosoritide.
Safety: Most common side effects include injection site reactions (redness, itching, or swelling) and a risk of low blood pressure (hypotension).
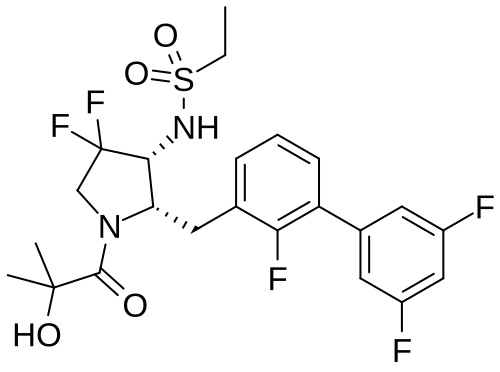
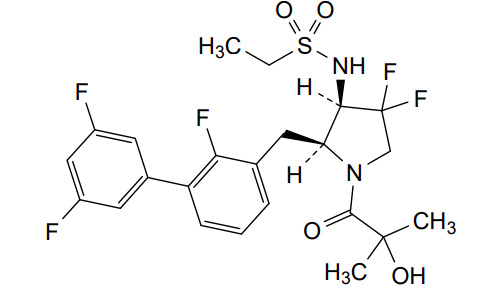
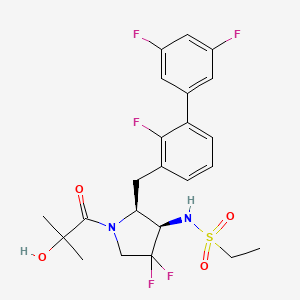
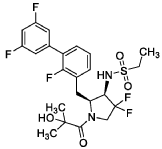
N-{(2S,3R)-4,4-difluoro-1-(2-hydroxy-2-methylpropanoyl)-2-[(2,3′,5′-trifluoro-[1,1′-biphenyl]-3-yl)methyl]pyrrolidin-3-yl}ethane-1-sulfonamide orexin type 2 receptor agonist, TAK-861, TAK 861, 59MF6P2ATF
Oveporexton is a small molecule drug. The usage of the INN stem ‘-orexton’ in the name indicates that Oveporexton is a orexin receptor agonist. Oveporexton has a monoisotopic molecular weight of 520.15 Da.
Oveporexton is being developed by Takeda.[1] As of July 2025, it has completed phase 3 clinical trials for treatment of narcolepsy, whereas no recent development has been reported for treatment of idiopathic hypersomnia.[1][5][10] Takeda plans to submit a New Drug Application (NDA) of oveporexton for the treatment of narcolepsy to the United StatesFood and Drug Administration (FDA) in 2025.[5] Oveporexton is a follow-on and replacement compound for Takeda’s earlier lead drug danavorexton (TAK-925), which is administered intravenously and stopped being developed due to unexpected liver toxicity findings.[10]
A Trial of TAK-861 for the Treatment of Narcolepsy With CataplexyCTID: NCT07363720Phase: Phase 3Status: Not yet recruitingDate: 2026-01-23Conditions: Narcolepsy Type 1 (NT1); Narcolepsy With CataplexyInterventions: PlaceboLinked Compound CID: 154617563
A Study of TAK-861 for the Treatment of Selected Central Hypersomnia ConditionsCTID: NCT05816382Phase: Phase 2/Phase 3Status: RecruitingDate: 2025-12-01Conditions: Narcolepsy Type 1Interventions: TAK-861Linked Compound CID: 154617563
A Study of TAK-861 in People With Narcolepsy Type 1CTID: NCT06505031Phase: Phase 3Status: CompletedDate: 2025-09-15Conditions: Narcolepsy Type 1Interventions: PlaceboLinked Compound CID: 154617563
A Study of TAK-861 for the Treatment of Narcolepsy Type 1CTID: NCT06470828Phase: Phase 3Status: CompletedDate: 2025-07-01Conditions: Narcolepsy Type 1Interventions: PlaceboLinked Compound CID: 154617563
A Study of TAK-861 in Participants With Narcolepsy Type 1CTID: NCT05687903Phase: Phase 2Status: CompletedDate: 2025-01-09Conditions: Narcolepsy Type 1Interventions: PlaceboLinked Compound CID: 154617563
A Randomized, Double-blind, Placebo-Controlled Study to Evaluate the Efficacy, Safety, and Tolerability of TAK-861 for the Treatment of Narcolepsy With Cataplexy (Narcolepsy Type 1)EudraCT: 2022-001654-38Phase: Phase 2Status: CompletedDate: 2023-05-26Linked Compound CID: 154617563
A Long-term Extension Study to Evaluate the Safety and Tolerability of TAK-861 in Participants With Selected Central Hypersomnia ConditionsEudraCT: 2022-002965-13Phase: Phase 2, Phase 3Status: Trial now transitionedDate: 2023-04-11Linked Compound CID: 154617563
A Randomized, Double-blind, Placebo-Controlled Study to Evaluate the Efficacy, Safety, and Tolerability of TAK-861 for the Treatment of Narcolepsy Without Cataplexy (Narcolepsy Type 2)EudraCT: 2022-002966-34Phase: Phase 2Status: CompletedDate: 2023-03-20Linked Compound CID: 154617563
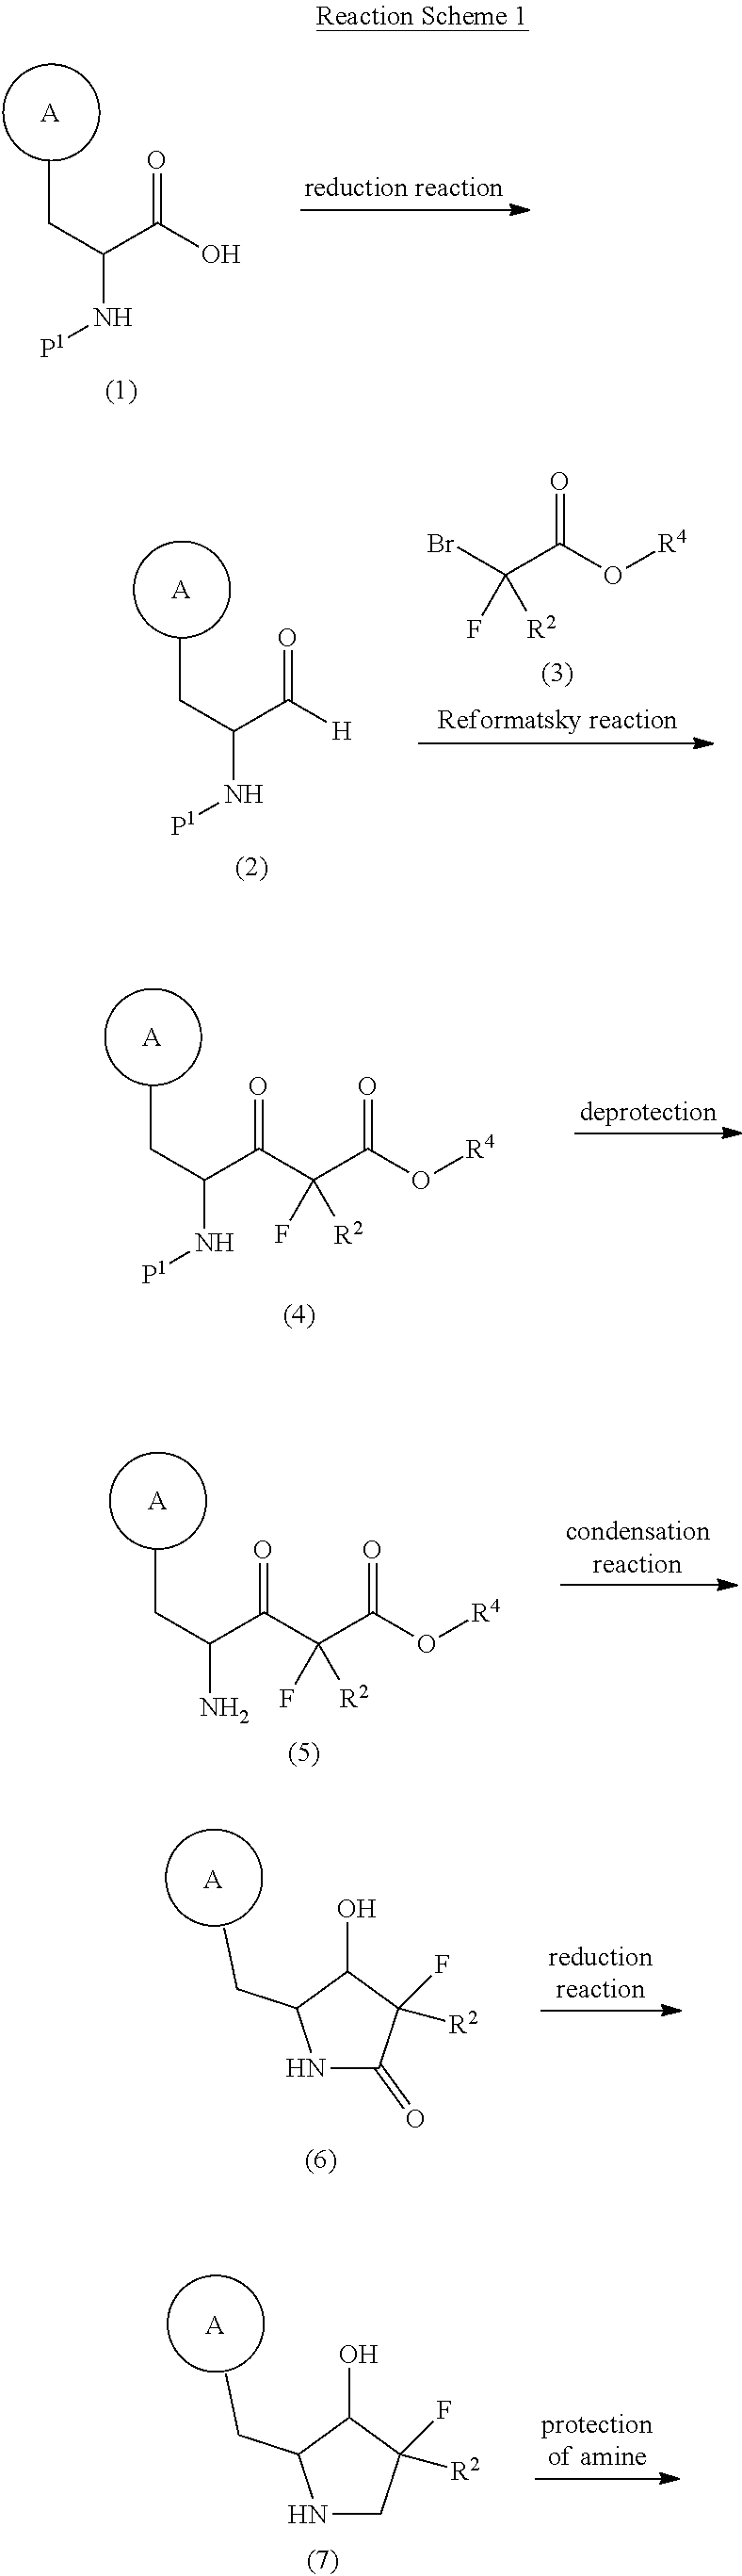
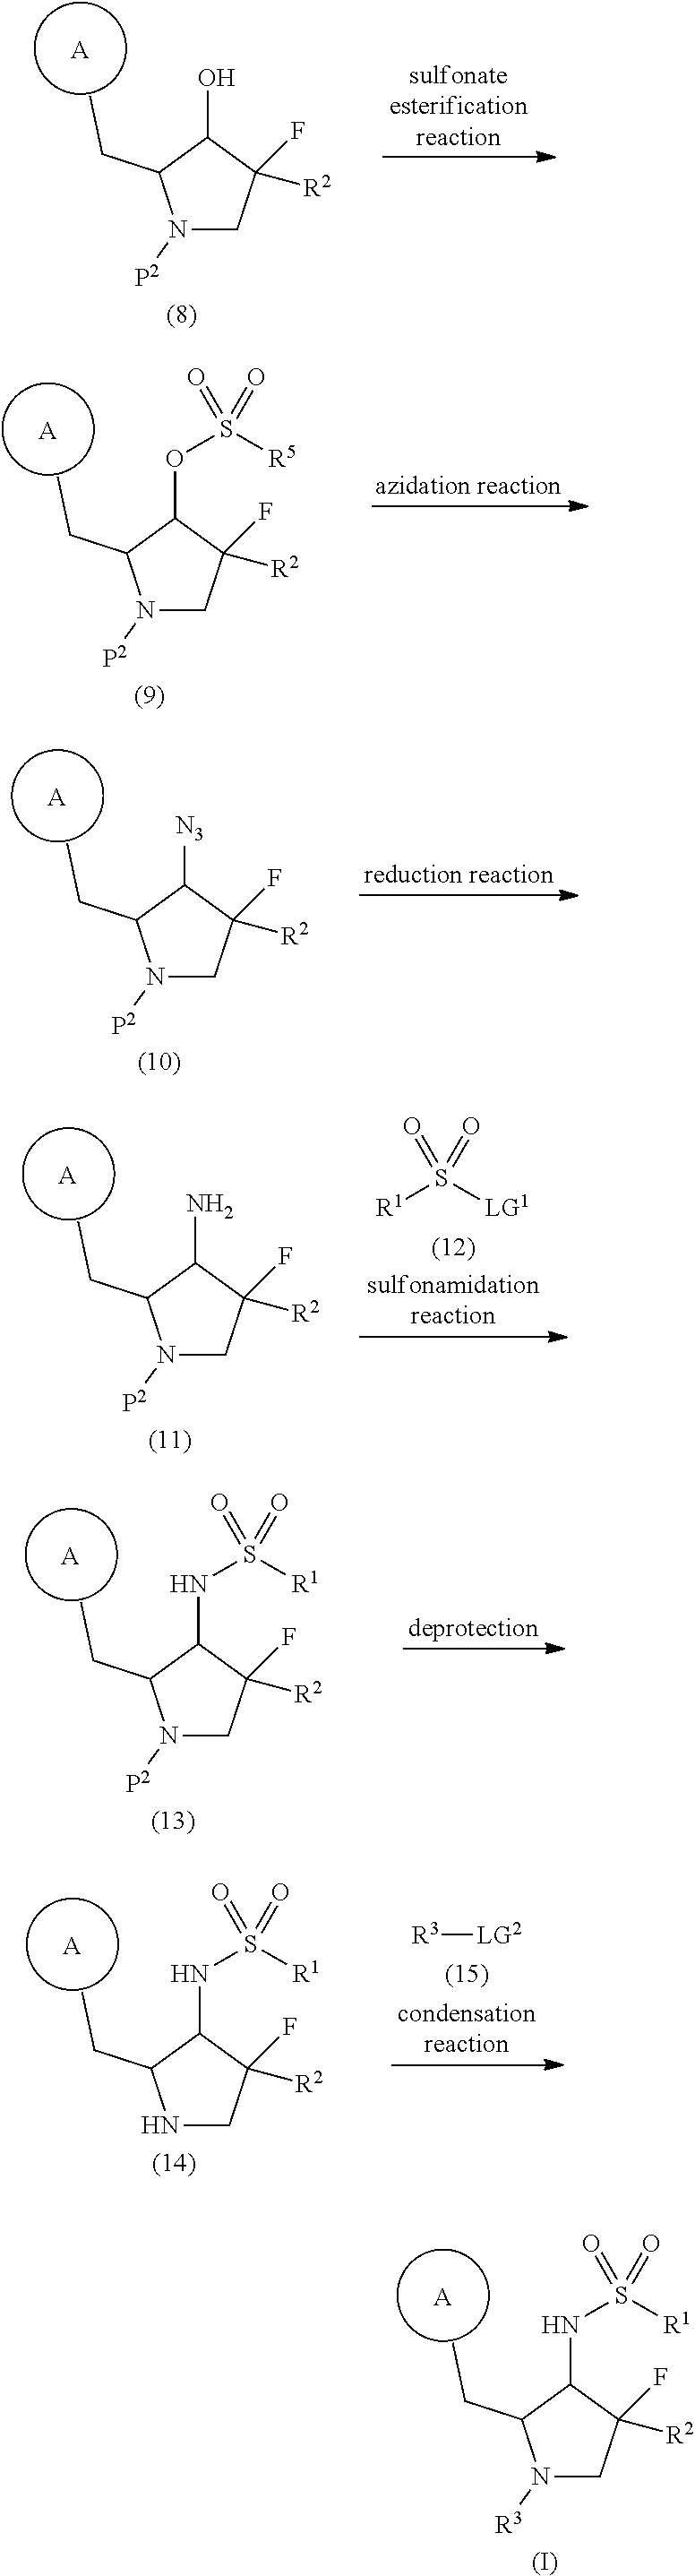
A) tert-butyl (2S,3R)-3-(ethylsulfonamido)-4,4-difluoro-2-((2,3′,5′-trifluoro-[1,1′-biphenyl]-3-yl)methyl)pyrrolidine-1-carboxylate
To a mixture of tert-butyl (2S,3R)-2-(3-chloro-2-fluorobenzyl)-3-(ethylsulfonamido)-4,4-difluoropyrrolidine-1-carboxylate (3.70 g), (3,5-difluorophenyl)boronic acid (2.56 g) and 1 M aqueous potassium phosphate solution (24.3 mL) in DME (50 mL) was added XPhos Pd G3 (0.343 g) at room temperature. The mixture was stirred at 90° C. under nitrogen atmosphere for 15 h. The reaction mixture was poured into water and extracted with EtOAc. The organic layer was washed with saturated brine, dried over magnesium sulfate and concentrated under reduced pressure. The residue was purified by column chromatography (silica gel, EOAc/hexane) to give the title compound (3.30 g).
MS: [M−H] − 533.2.
B) N-((2S,3R)-4,4-difluoro-2-((2,3′,5′-trifluoro-[1,1′-biphenyl]-3-yl)methyl)pyrrolidin-3-yl)ethanesulfonamide hydrochloride
A mixture of tert-butyl (2S,3R)-3-(ethylsulfonamido)-4,4-difluoro-2-((2,3′,5′-trifluoro-[1,1′-biphenyl]-3-yl)methyl)pyrrolidine-1-carboxylate (3.30 g) and 4 M HCl/CPME solution (30 mL) was stirred overnight at room temperature. By filtration, the title compound (2.86 g) was obtained.
MS: [M+H] + 435.1.
C) N-{(2S,3R)-4,4-difluoro-1-(2-hydroxy-2-methylpropanoyl)-2-[(2,3′,5′-trifluoro[1,1′-biphenyl]-3-yl)methyl]pyrrolidin-3-yl}ethanesulfonamide
To a mixture of N-((2S,3R)-4,4-difluoro-2-((2,3′,5′-trifluoro-[1,1′-biphenyl]-3-yl)methyl)pyrrolidin-3-yl)ethanesulfonamide hydrochloride (200 mg) and DIPEA (0.367 ml) in THF (3 mL) was added alpha-acetoxy-isobutyryl chloride (0.074 ml) at 0° C., and the mixture was stirred at same temperature for 10 min. To the mixture were added water (1 ml) and 4 M lithium hydroxide solution (1.06 ml), and the mixture was stirred overnight at room temperature. The mixture was diluted with saturated brine and extracted with EtOAc. The extract was dried over magnesium sulfate and concentrated under reduced pressure. The residue was purified by column chromatography (silica gel, EtOAc/hexane) and recrystallized from EtOAc/hexane to give the title compound (154 mg).
1H NMR (400 MHz, CDCl 3) δ 1.32-1.40 (9H, m), 2.27-2.54 (1H, m), 2.88-3.16 (4H, m), 4.02-4.49 (3H, m), 4.86-5.20 (2H, m), 6.78-6.86 (1H, m), 7.02-7.10 (2H, m), 7.16-7.22 (1H, m), 7.27-7.31 (1H, m), 7.35-7.43 (1H, m).
N-{(2S,3R)-4,4-difluoro-l-(2-hydroxy-2-methylpropanoyl)-2-[(2,3′,5′-trifluoro[l,l’-biphenyl]-3-yl)methyl]pyrrolidin-3-yl}ethanesulfonamide, (hereafter referred as to “Compound A”) is described in U.S. Patent No. 11,028,048.
Dauvilliers Y, Plazzi G, Mignot E, Lammers GJ, Del Río Villegas R, Khatami R, et al. (May 2025). “Oveporexton, an Oral Orexin Receptor 2-Selective Agonist, in Narcolepsy Type 1”. The New England Journal of Medicine. 392 (19): 1905–1916. doi:10.1056/NEJMoa2405847. PMID40367374.
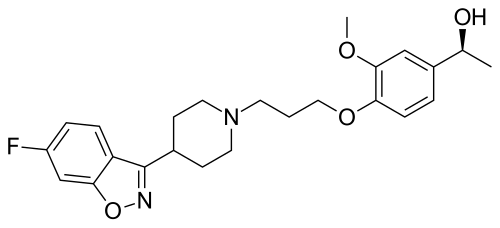
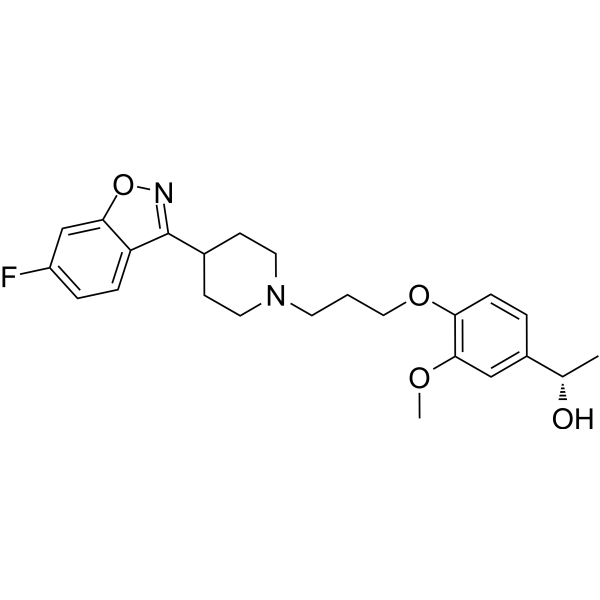
25 Feb 2026Vanda Pharmaceuticals has patent protection for an improved method of treatment with milsaperidone in USA
25 Feb 2026Vanda Pharmaceuticals has patents pending for an improved method of treatment with milsaperidone in China, Australia, Israel, Mexico and worldwide
56.36 g of boran complex of (3aR, 7R)-1-methyl-3,3-diphenyl-tetrahydro-pyrrolo[1,2-c][1 ,3,2]oxazaborole (1 equivalent) is dissolved under nitrogen in methylenchloride, and the solution is cooled to 0°C. A 1M solution of 1-(4-{3-[4-(6-fluoro-benzo[d]isoxazol-3-yl)-piperidin-1-yl]-propoxy}-3-methoxy-phenyl)-ethanone (iloperidone; 1 equivalent) in methylenchloride is added via a dropping funnel over 90 minutes while the internal temperature is maintained at 0°C ± 2°C. After the addition is complete, the mixture is stirred at 0°C for 20 hours. The reaction mixture is then poured into precooled methanol (0-5°C) during 1 hour. The solution is warmed to room temperature and stirred until the H2 evolution ceases. The solution is concentrated by distillation and the residue dried in vacuum, treated with methanol and stirred for about 1 hour at 50°C and an additional hour at 0CC. The product is isolated by filtration and dried under reduced pressure for 3 hours at 50°C. The title compound is obtained (white crystals).
[α]D20– 19.3° (c=1 in chloroform) Mp: 138.2 – 138.8°C
The boran complex used as starting material can be obtained as follows:
200 ml of a solution of (3aR, 7R)-1-methyl-3,3-diphenyl-tetrahydro-pyrrolo[1,2-c][1,3,2]oxazaborole (1M in toluene) is stirred at room temperature under nitrogen. 1.2 equivalent borane-dimethylsulfide complex is added with a syringe. The solution is stirred for 2 further hours at room temperature. The borane complex is then crystallised by addition of 4 vol dry hexane and cooling to -12°C for 1.5 hour. The product is isolated by filtration in a sintered glass funnel and dried in vacuum at 40°C. The boran complex is obtained /white crystals).
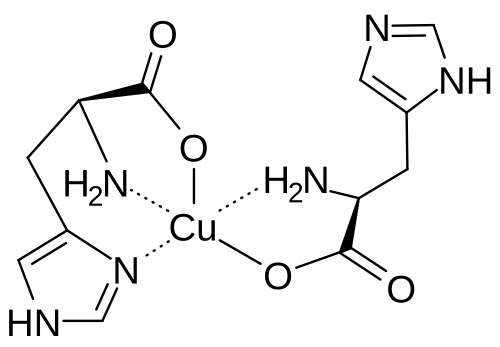
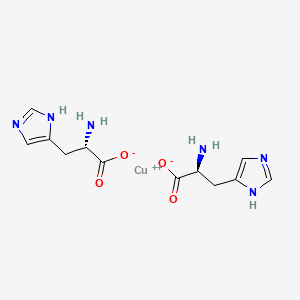
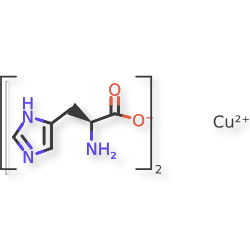
Copper histidinate, sold under the brand name Zycubo, is a medication used for the treatment of Menkes disease.[1] Copper histidinate is a copper replacement therapy given by subcutaneous injection.[1][2]
The most common side effects include infections, respiratory problems, seizures, vomiting, fever, anemia and injection site reactions.[2]
Copper histidinate was approved for medical use in the United States in January 2026.[2]
Menkes disease is a neurodegenerative disorder caused by a genetic defect that impairs a child’s ability to absorb copper.[2] The disease is characterized by seizures, failure to gain weight and grow, developmental delays, and intellectual disability.[2] It leads to abnormalities of the vascular system, bladder, bowel, bones, muscles, and nervous system.[2]
SYN
A275388 — Flores-Pulido AA, Jimenez-Perez VM, Garcia-Chong NR: Sintesis y uso de histidinato de cobre en ninos con enfermedad de Menkes en Mexico. Gac Med Mex. 2019;155(2):191-195. doi: 10.24875/GMM.18004310. [PubMed:31056589]
World Health Organization (2025). “International nonproprietary names for pharmaceutical substances (INN): recommended INN: list 94”. WHO Drug Information. 39 (3). hdl:10665/383022.
Clinical trial number NCT00811785 for “Molecular Bases of Response to Copper Treatment in Menkes Disease, Related Phenotypes, and Unexplained Copper Deficiency” at ClinicalTrials.gov
Mechanism of ActionSodium-glucose transporter 2 inhibitors
PreregistrationType 2 diabetes mellitus
04 Sep 2025Chemical structure information added.
31 Dec 2023Preregistration for Type 2 diabetes mellitus in China (PO), in December 2023
31 Dec 2023Efficacy and adverse events data from a phase IIIa trial in Type 2 diabetes mellitus released by Sunshine Lake Pharma, before December 2023
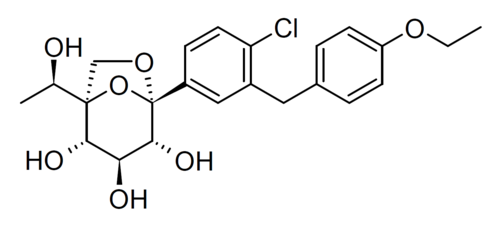
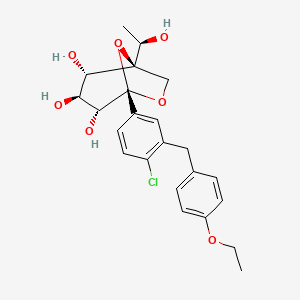
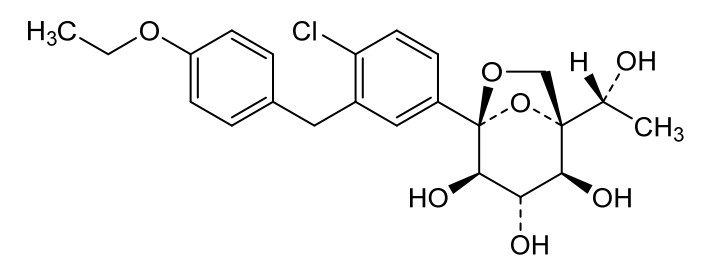
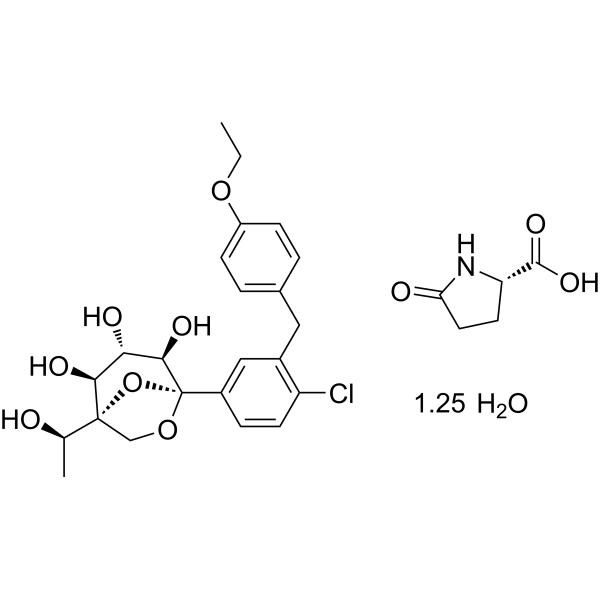
HEC Pharma announced that its independently developed Olorigliflozin (previously known as rongliflozin pyroglutamate) capsules, has been approved for marketing by the China NMPA. It can be used as a monotherapy or in combination with metformin to improve glycemic control in adults with T2DM.
Olorigliflozin is a SGLT2 inhibitor that moderately inhibits SGLT1. It promotes urinary glucose excretion by potently inhibiting renal SGLT2 receptors and reduces glucose or galactose absorption by moderately inhibiting intestinal SGLT1 receptors, effectively reducing drastic postprandial blood glucose fluctuations.
Rongliflozin (DJT1116PG) is a selective and orally active inhibitor of sodium-glucose co-transporter-2 (SGLT-2). Rongliflozin can be used for the research of type 2 diabetes mellitus (T2DM).
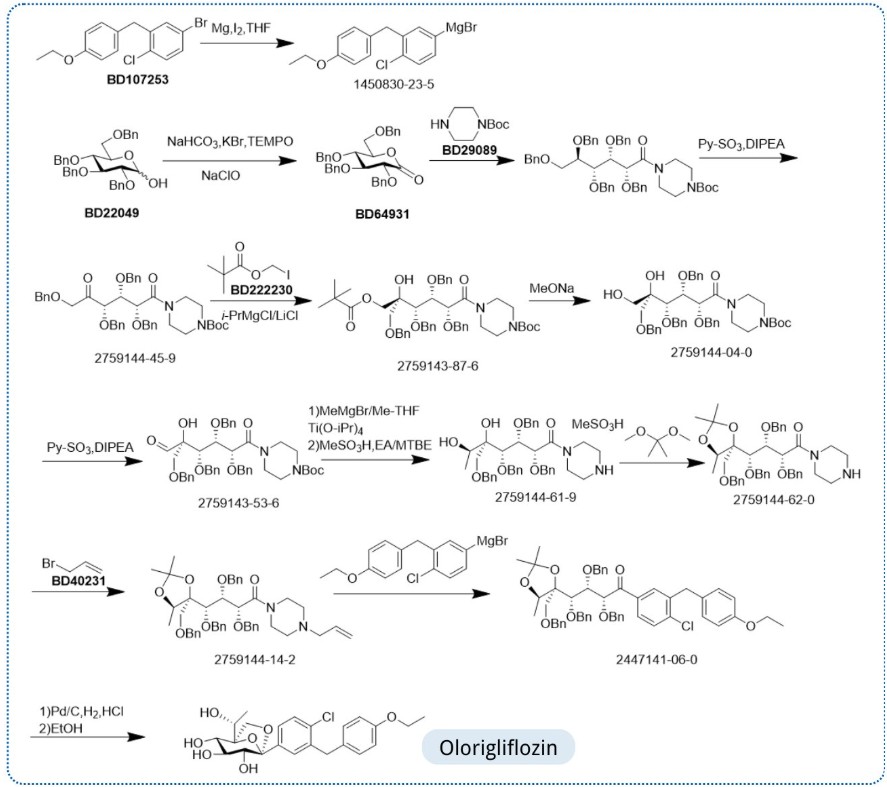
On January 16, 2026, Olorigliflozin, a small-molecule antidiabetic drug was approved by the National Medical Products Administration (NMPA) of China for improving glycemic control in adult patients with type 2 diabetes mellitus. Its synthetic route highlights a key and recurring feature of SGLT inhibitors
a glucose-derived core, as a sugar ring is present in the majority of approved SGLT2 inhibitors and serves as the structural foundation of these molecules.
#BLDSynthesis 🧪 From Approved Olorigliflozin to Glucose-Derived Building Blocks
On January 16, 2026, Olorigliflozin, a small-molecule antidiabetic drug was approved by the National Medical Products Administration (NMPA) of China for improving glycemic control in adult patients… pic.twitter.com/B7R5rePJ3H
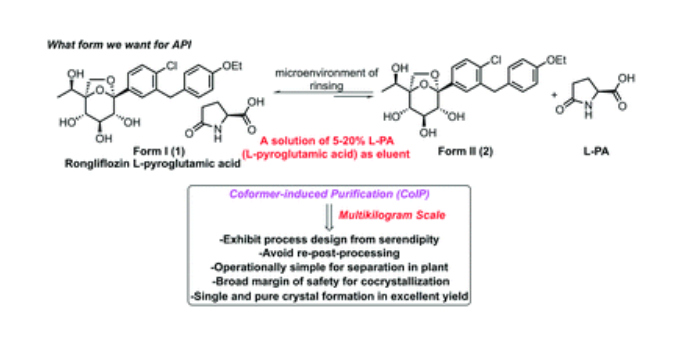
Rongliflozin L-pyroglutamic acid, a highly active SGLT-2 inhibitor cocrystal discovered and developed by our group, is currently undergoing clinical trials for the treatment of diabetes. Here, we report and design a simple and robust process to obtain a single and pure crystalline form I (1) of the cocrystal, containing Rongliflozin (2) with L-pyroglutamic acid (L-PA), based on coformer-induced purification (CoIP). Extensive experiments showed that the addition of L-pyroglutamic acid in the eluent was key to suppression of the dissociation equilibrium of the cocrystal during lessivation, with high efficiency. Importantly, based in this profile, this process exhibited strong robustness and margin of safety at multigram and multikilogram scales
Kilogram scale Process of 1
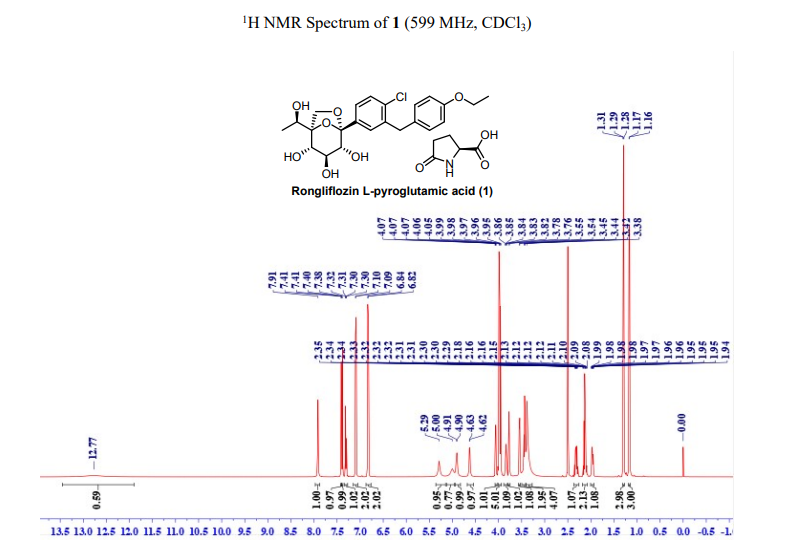
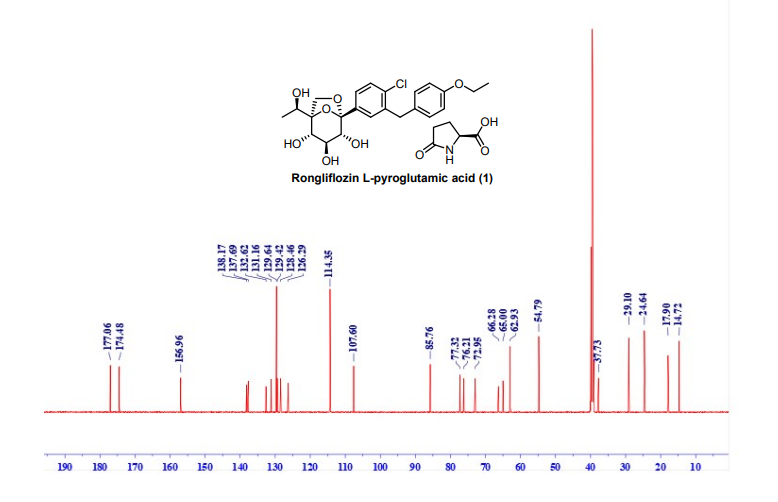
A mixture of (1R,2S,3S,4R,5S)-5-(4-chloro-3-(4-ethoxybenzyl) phenyl)-1-((R)-1- hydroxyethyl)-6,8-dioxabicyclo [3.2.1] octane-2,3,4-triol ethanolate form III (3) (23.45 kg, 47.3 mol), L-pyroglutamic acid (24.31 kg, 4.0 equiv.), EtOH (35.9 L) and H2O (70 L) was added into a 300 L reactor at room temperature. The slurry was heated to 65 °C and stirred until it is clear. The clear solution was cooled to 35±5 °C typically. Seed crystal form I (1) (0.70 kg, 3% g/g) was added when the solution was cooled to 34 °C and maintained for 1.5 h. Gradually, the slurry was cool to 30 °C and 25 °C in 3 hours, and finally stirred at 25 °C for 24 h. The slurry was collected on a centrifuge filter. The filter cake was washed with a mixed solution of EtOH (31.3 L)/H2O (62.7 L) with L-pyroglutamic acid (1.64 kg, 7% g/g) pre-cooled to -15°C. The wet cake was dried under vacuum at 45 °C for 8 h. Pure cocrystal form I (1) was obtained as a white solid (24.91 kg, yield 91%). MP (DSC onset) = 96.91 ℃. 1H NMR (599 MHz, DMSO-d6) δ 12.77 (br, 1H), 7.91 (s, 1H), 7.41 (d, J = 2.0 Hz, 1H), 7.39 (d, J = 12.0 Hz, 1H), 7.31 (dd, J = 12.0, 2.0 Hz, 1H), 7.10 (d, J = 2.0 Hz , 2H), 6.83 (d, J = 2.0 Hz, 2H), 5.29 (s, 1H), 5.00 (s, 1H), 4.91 (d, J = 6.7 Hz, 1H), 4.63 (d, J = 6.1 Hz, 1H), 4.06 (dd, J = 12.0, 6.0 Hz, 1H), 3.99– 3.95 (m, 5H), 3.84 (p, J = 6.0 Hz, 1H), 3.77 (d, J = 12.0 Hz, 1H), 3.55 (d, J = 6.0 Hz, 1H), 3.44 (t, J = 12.0 Hz, 2H), 3.38 (s, 4H), 2.35-2.29 (m, 1H), 2.18-2.08 (m, 2), 1.99-1.94 (m, 1H), 1.29 (t, J = 12.0 Hz, 3H), 1.17 (d, J = 6.0 Hz, 3H). 13C NMR (151 MHz, DMSO-d6) δ 177.06, 174.48, 156.96, 138.17, 137.69, 131.16, 129.64, 129.42, 128.46, 126.29, 114.35, 107.60, 85.76, 77.32, 76.21, 72.95, 66.28, 65.00, 62.93, 54.79, 37.73, 29.10, 24.64, 17.90, 14.72. HRMS: (ESI) Calcd for C23H27ClO7 [M+NH4]+: 468.1784, C5H7NO3 [M+H]+ :130.0499; Found: 468.1774, 130.0490 respectively. IR (KBr, cm-1): 3257, 2986, 2927, 1750, 1648, 1513, 1476, 1371, 1264, 1239, 1223, 1206, 1088, 1061, 821
13C NMR
AS ON JUNE2025 4.45 LAKHS VIEWS ON BLOG WORLDREACH AVAILABLEFOR YOUR ADVERTISEMENT
Zhang H, Liu J, Zhu X, Li X, Chen H, Wu M, et al. (May 2020). “A Phase I Study on the Pharmacokinetics and Pharmacodynamics of DJT1116PG, a Novel Selective Inhibitor of Sodium-glucose Cotransporter Type 2, in Healthy Individuals at Steady State”. Clinical Therapeutics. 42 (5): 892–905.e3. doi:10.1016/j.clinthera.2020.03.007. PMID32265061.
Zhang H, Zhu X, Li X, Chen H, Wu M, Li C, et al. (February 2020). “Pharmacokinetics and pharmacodynamics of rongliflozin, a novel selective inhibitor of sodium-glucose co-transporter-2, in people with type 2 diabetes mellitus”. Diabetes, Obesity & Metabolism. 22 (2): 191–202. doi:10.1111/dom.13887. PMID31588657.
Baxdrostat is under investigation in clinical trial NCT06344104 (A Phase III Study to Investigate the Efficacy and Safety of Baxdrostat in Asian Participants With Uncontrolled Hypertension on Two or More Medications Including Participants With Resistant Hypertension).
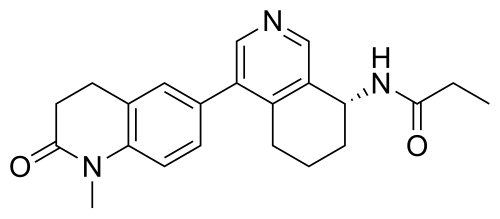
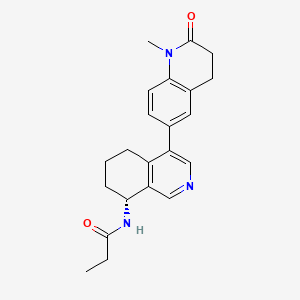
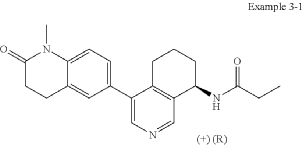
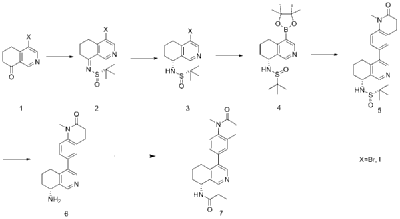
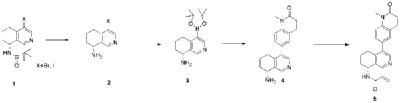
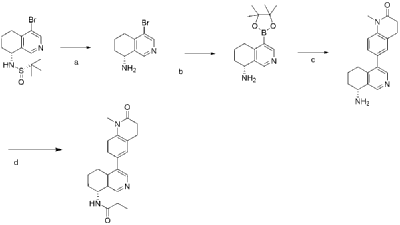
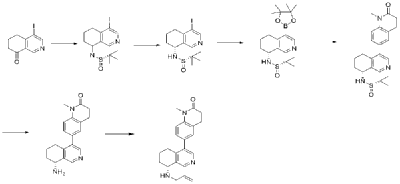
In analogy to the procedures described for the preparation of intermediate A-2 [E] and for the preparation of intermediate B-1, Suzuki reaction of (+)-(R)-4-bromo-5,6,7,8-tetrahydroisoquinolin-8-amine (intermediate B-3b) with 1-methyl-6-(4,4,5,5-tetramethyl-[1,3,2]dioxaborolan-2-yl)-3,4-dihydro-1H-quinolin-2-one (intermediate A-1) gave (R)-6-(8-amino-5,6,7,8-tetrahydroisoquinolin-4-yl)-1-methyl-3,4-dihydroquinolin-2(1H)-one and after subsequent reaction with propionyl chloride the title compound as colorless solid. MS: 364.2 (M+H +).
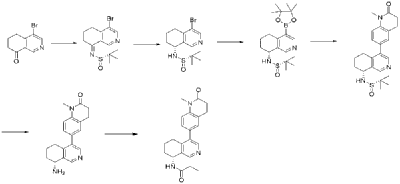
Dissolve 4-bromo-6,7-dihydroisoquinolin-8(5H)-one (1.56 g, 6.9 mmol) and (S)-tert-butylsulfenamide (2.51 g, 20.7 mmol) in 20 mL of tetrahydrofuran. Add ethyl titanate (10.08 mL, 48.28 mmol). Heat to 65°C and stir for 48 hours. Cool to room temperature, add ethyl acetate and water, stir for 15 minutes, and remove the resulting solid by filtration. Separate the liquids, dry the organic phase over anhydrous sodium sulfate, filter, and evaporate to dryness under reduced pressure to obtain the crude product (S,Z)-N-(4-bromo-6,7-dihydroisoquinolin-8(5H)-tert-butylsulfenimide), which is used directly in the next step.
Step B
Compound (S,Z)-N-(4-bromo-6,7-dihydroisoquinoline-8(5H)-tert-butylsulfonyl imide) (1.98 g, 6 mmol) was dissolved in 15 mL of tetrahydrofuran and cooled to -45°C. Sodium borohydride (0.34 g, 9.0 mmol) was added, and the mixture was allowed to return to room temperature and stirred for 18 hours. The mixture was quenched with ice water and extracted with dichloromethane. The resulting organic phase was washed with saturated brine, dried over anhydrous sodium sulfate, and evaporated to dryness under reduced pressure. The residue was purified by column chromatography to obtain compound (S)-N-(4-bromo-6,7-dihydroisoquinoline-8(5H))-tert-butylsulfonyl imide (755 mg, 38% yield). LC/MS (ESI): m/z = 331.2 [M+H] + .
Step C
To a mixture of (S)-N-(4-bromo-6,7-dihydroisoquinoline-8(5H))-tert-butylsulfonimide (0.66 g, 2 mmol), pinacol diboronate (1.05 g, 2.1 mmol), and AcOK (0.578 g, 6 mmol) in toluene (10 mL) was added Pd(dppf)Cl 2 (0.144 g, 0.2 mmol). The mixture was degassed and stirred at 130 ° C for 3 hours. The reaction mixture was filtered and concentrated to give a residue. EtOAc (15 mL) and water (10 mL) were added to the residue. The organic phase was washed with brine (50 mL), dried over anhydrous sodium sulfate, filtered and concentrated to give a residue. The residue was purified by column chromatography (SiO 2 ) and eluted with 30-40% ethyl acetate in petroleum ether to afford (S)-N-tert-butylsulfonamido-6,7-dihydroisoquinolin-8(5H)-4-boronic acid pinacol ester (0.45 g, 60% yield). LC/MS (ESI): m/z = 378.3 [M+H] + .
Step D
To a reaction flask, add 6-bromo-1-methyl-3,4-dihydroquinolin-2(1H)-one (0.29 g, 1.2 mmol), (S)-N-tert-butylsulfonamido-6,7-dihydroisoquinolin-8(5H)-4-boronic acid pinacol ester (0.42 g, 1.26 mmol), bistriphenylphosphine palladium dichloride (84 mg, 0.12 mmol), cuprous iodide (38 mg, 0.2 mmol), triethylamine (1.01 g, 10.0 mmol), and 15 mL of N,N-dimethylformamide. The atmosphere was purged with nitrogen three times and the reaction was stirred at 90°C overnight. After cooling to room temperature, the reaction mixture was diluted with ethyl acetate and water, and extracted with ethyl acetate. The resulting organic phase was washed with water and saturated brine, dried over anhydrous sodium sulfate, and evaporated to dryness under reduced pressure. The residue was purified by column chromatography to afford (S)-2-methyl-N-((R)-4-(1-methyl-2-oxo-1,2,3,4-tetrahydroquinolin-6-yl)-5,6,7,8-tetrahydroisoquinolin-8-yl)tert-butylsulfonimide (0.37 g, 74% yield) as a yellow solid. LC/MS (ESI): m/z = 411.5 [M+H] + .
Step E
Compound (S)-2-methyl-N-((R)-4-(1-methyl-2-oxo-1,2,3,4-tetrahydroquinolin-6-yl)-5,6,7,8-tetrahydroisoquinolin-8-yl)tert-butylsulfonimide (0.33 g, 0.80 mmol) was dissolved in 1 mL of dichloromethane, and 1 mL of trifluoroacetic acid was added. The mixture was stirred and reacted for 1 hour. The reaction solution was concentrated under reduced pressure. The residue was purified by reverse preparative column chromatography to obtain compound (R)-6-(8-amino-5,6,7,8-tetrahydroisoquinolin-4-yl)-1-methyl-3,4-dihydroquinolin-2(1H)-one (0.24 g, 97% yield). LC/MS (ESI): m/z = 307.1 [M+H] + .
Step F
To a reaction flask, add (R)-6-(8-amino-5,6,7,8-tetrahydroisoquinolin-4-yl)-1-methyl-3,4-dihydroquinolin-2(1H)-one (100 mg, 0.33 mmol), triethylamine (51 mg, 0.5 mmol), and 4 ml of tetrahydrofuran. After cooling in an ice-water bath, slowly add a solution of propionyl chloride (46.25 mg, 0.5 mmol) in 0.5 ml of tetrahydrofuran dropwise. Stirring is continued for 4 hours after addition. The reaction mixture is quenched with methanol and evaporated to dryness under reduced pressure. The residue is purified by column chromatography to obtain the target compound, Baxdrostat (46 mg, 38% yield). LC/MS(ESI):m/z=363.1[M+H]+.H NMR(400MHz, CDCl3)ppm 1.22(t,3H)1.79(s,3H)2.07(s,1H)2.28(q,2H)2.43-2.68(m,2H)2.71(t,2H)2.82-3.12(m,2H) 3.40(s,3H)5.34(d,1H)5.78(d,1H)7.05(d,1H)7.09(s,1H)7.17(d,1H)8.28(s,1H)8.49(s,1H)
Example 2
Step A
Compound (S)-N-(4-bromo-6,7-dihydroisoquinolin-8(5H))-tert-butylsulfonylimide (1.65 g, 5 mmol) was dissolved in 20 mL of dichloromethane, and 20 mL of trifluoroacetic acid was added. The mixture was stirred and reacted for 1 hour. The reaction solution was concentrated under reduced pressure. The residue was purified by reverse-phase preparative column chromatography to obtain compound (R)-4-bromo-5,6,7,8-tetrahydroisoquinolin-8-amine (1.07 g, 94% yield). LC/MS (ESI): m/z = 226.0 [M+H] + .
Step B
To a mixture of (R)-4-bromo-5,6,7,8-tetrahydroisoquinolin-8-amine (0.86 g, 3.8 mmol), pinacol diboron (2 g, 4 mmol), AcOK (1.10 g, 11.4 mmol) in toluene (10 mL) was added Pd(dppf)Cl 2 (0.27 g, 0.38 mmol). The mixture was degassed and stirred at 130 ° C for 3 hours. The reaction mixture was filtered and concentrated to give a residue. EtOAc (10 mL) and water (10 mL) were added to the residue. The organic phase was washed with brine (10 mL), dried over anhydrous sodium sulfate, filtered and concentrated to give a residue. The residue was purified by column chromatography (SiO 2 ) and eluted with 30-40% ethyl acetate in petroleum ether to afford (R)-8-amino-5,6,7,8-tetrahydroisoquinoline-4-boronic acid pinacol ester (0.68 g, 65% yield). LC/MS (ESI): m/z = 274.1 [M+H] + .
Step C
To a reaction flask, add 6-bromo-1-methyl-3,4-dihydroquinolin-2(1H)-one (0.72 g, 3.0 mmol), (R)-8-amino-5,6,7,8-tetrahydroisoquinolin-4-boronic acid pinacol ester (0.99 g, 3.6 mmol), bistriphenylphosphine palladium dichloride (210 mg, 0.3 mmol), and potassium phosphate monohydrate (204 mg, 0.9 mmol). Dissolve the mixture in dioxane and water (9:1, 30 mL). Replace the atmosphere with nitrogen three times and allow the mixture to react overnight at 90°C with stirring. Cool to room temperature, dilute the reaction solution with ethyl acetate and water, and extract with ethyl acetate. The resulting organic phase is then washed with water and saturated brine, dried over anhydrous sodium sulfate, and evaporated to dryness under reduced pressure. The residue was purified by column chromatography to obtain (R)-6-(8-amino-5,6,7,8-tetrahydroisoquinolin-4-yl)-1-methyl-3,4-dihydroquinolin-2(1H)-one (0.81 g, 88% yield). LC/MS (ESI): m/z = 307.1 [M+H] + . The target compound, Baxdrostat, was then prepared using a method similar to the last step in Example 1.
Example 3
Step A
4-Bromo-6,7-dihydroisoquinolin-8(5H)-one (1.88 g, 6.9 mmol) and (S)-tert-butylsulfenamide (2.51 g, 20.7 mmol) were dissolved in 20 mL of tetrahydrofuran. Ethyl titanate (10.08 mL, 48.28 mmol) was added and the mixture was heated to 65°C with stirring for 48 hours. After cooling to room temperature, ethyl acetate and water were added and stirred for 15 minutes. The resulting solid was removed by filtration. The organic phase was separated and dried over anhydrous sodium sulfate, filtered, and evaporated to dryness under reduced pressure to obtain the crude product (S,Z)-N-(4-bromo-6,7-dihydroisoquinolin-8(5H)-tert-butylsulfenimide), which was used directly in the next step. LC/MS (ESI): m/z = 376.2 [M+H] + .
Step B
Compound (S,Z)-N-(4-iodo-6,7-dihydroisoquinoline-8(5H)-tert-butylsulfonyl imide) (2.26 g, 6 mmol) was dissolved in 15 mL of tetrahydrofuran and cooled to -45°C. Sodium borohydride (0.36 g, 9.0 mmol) was added, and the mixture was allowed to return to room temperature and stirred for 18 hours. The mixture was quenched with ice water and extracted with dichloromethane. The resulting organic phase was washed with saturated brine, dried over anhydrous sodium sulfate, and evaporated to dryness under reduced pressure. The residue was purified by column chromatography to obtain compound (S)-N-(4-iodo-6,7-dihydroisoquinoline-8(5H))-tert-butylsulfonyl imide (1.04 g, 46% yield). LC/MS (ESI): m/z = 378.0 [M+H] + .
Step C
To a mixture of (S)-N-(4-iodo-6,7-dihydroisoquinoline-8(5H))-tert-butylsulfonimide (0.76 g, 2 mmol), pinacol diboronate (1.05 g, 2.1 mmol), and AcOK (0.578 g, 6 mmol) in toluene (10 mL) was added Pd(dppf)Cl 2 (0.144 g, 0.2 mmol). The mixture was degassed and stirred at 130 ° C for 3 hours. The reaction mixture was filtered and concentrated to give a residue. EtOAc (15 mL) and water (10 mL) were added to the residue. The organic phase was washed with brine (50 mL), dried over anhydrous sodium sulfate, filtered and concentrated to give a residue. The residue was purified by column chromatography (SiO 2 ) and eluted with 30-40% ethyl acetate in petroleum ether to afford (S)-N-tert-butylsulfonamido-6,7-dihydroisoquinolin-8(5H)-4-boronic acid pinacol ester (0.51 g, 68% yield). LC/MS (ESI): m/z = 378.2 [M+H] + .
The next three steps were carried out in the same manner as in Example 1 to prepare the target compound Baxdrostat.
A new treatment has been shown to significantly lower blood pressure in people whose levels stay dangerously high, despite taking several existing medicines, according to the results of a Phase III clinical trial led by a UCL Professor. Globally, around 1.3 billion people have high blood pressure (hypertension), and in around half of cases the condition is uncontrolled or treatment resistant. These individuals face a much greater risk of heart attack, stroke, kidney disease, and early death. In the UK the number of people with hypertension is around 14 million.
The international BaxHTN trial, led by Professor Bryan Williams (UCL Institute of Cardiovascular Science), assessed the new drug baxdrostat—which is taken as a tablet—with participation from nearly 800 patients across 214 clinics worldwide.
The trial results showed that, after 12 weeks, patients taking baxdrostat (1 mg or 2 mg once daily in pill form) saw their blood pressure fall by around 9-10 mmHg more than placebo—a reduction large enough to cut cardiovascular risk. About four in 10 patients reached healthy blood pressure levels, compared with fewer than two in 10 on placebo.
Principal Investigator, Professor Williams, who is presenting the results at ESC, said, “Achieving a nearly 10 mmHg reduction in systolic blood pressure with baxdrostat in the BaxHTN Phase III trial is exciting, as this level of reduction is linked to substantially lower risk of heart attack, stroke, heart failure and kidney disease.”
How baxdrostat works
Blood pressure is strongly influenced by a hormone called aldosterone, which helps the kidneys regulate salt and water balance.
Some people produce too much aldosterone, causing the body to hold onto salt and water. This aldosterone dysregulation pushes blood pressure up and makes it very difficult to control.
Addressing aldosterone dysregulation has been a key effort in research over many decades, but it has been so far difficult to achieve.
Baxdrostat works by blocking aldosterone production, directly addressing this driver of high blood pressure (hypertension).
Professor Williams, Chair of Medicine at UCL, said, “These findings are an important advance in treatment and in our understanding of the cause of difficult-to-control blood pressure.
“Around half of people treated for hypertension do not have it controlled, however this is a conservative estimate and the number is likely higher, especially as the target blood pressure we try to reach is now much lower than it was previously.
“In patients with uncontrolled or resistant hypertension, the addition of baxdrostat 1mg or 2mg once daily to background antihypertensive therapy led to clinically meaningful reductions in systolic blood pressure, which persisted for up to 32 weeks with no unanticipated safety findings.
“This suggests that aldosterone is playing an important role in causing difficult to control blood pressure in millions of patients and offers hope for more effective treatment in the future.”
Historically, higher-income Western countries were reported to have far higher levels of hypertension. However, largely due to changing diets (adding less salt to food), the numbers of people living with the condition is now far higher in Eastern and lower-income countries. More than half of those affected live in Asia, including 226 million people in China and 199 million in India.
Professor Williams added, “The results suggest that this drug could potentially help up to half a billion people globally—and as many as 10 million people in the UK alone, especially at the new target level for optimal blood pressure control.”
AS ON JUNE2025 4.45 LAKHS VIEWS ON BLOG WORLDREACH AVAILABLEFOR YOUR ADVERTISEMENT
Dogra S, Shah S, Gitzel L, Pusukur B, Sood A, Vyas AV, Gupta R (July 2023). “Baxdrostat: A Novel Aldosterone Synthase Inhibitor for Treatment Resistant Hypertension”. Current Problems in Cardiology. 48 (11): 101918. doi:10.1016/j.cpcardiol.2023.101918. PMID37399857. S2CID259320969.
Awosika A, Cho Y, Bose U, Omole AE, Adabanya U (October 2023). “Evaluating phase II results of Baxdrostat, an aldosterone synthase inhibitor for hypertension”. Expert Opinion on Investigational Drugs. 32 (11): 985–995. doi:10.1080/13543784.2023.2276755. PMID37883217. S2CID264517675.
To treat platinum-resistant epithelial ovarian, fallopian tube, or primary peritoneal cancer after one to three prior systemic treatment regimens, at least one of which included bevacizumab
Mechanism of ActionGlucocorticoid receptor antagonists
Orphan Drug StatusYes – Pancreatic cancer; Cushing syndrome
Phase IIICushing syndrome; Ovarian cancer; Pancreatic cancer
Phase IIFallopian tube cancer; Peritoneal cancer; Prostate cancer
Phase I/IISolid tumours
Phase IAdrenocortical carcinoma
Most Recent Events
09 Sep 2022Subgroup analysis efficacy data from a phase-II trial in Ovarian cancer presented at the 47th European Society for Medical Oncology Congress (ESMO-2022)
29 Jun 2022Phase-III clinical trials in Ovarian cancer (Combination therapy, Recurrent, Second-line therapy or greater) in USA (PO)
06 Jun 2022Corcept Therapeutics announces intentions to submit a NDA for Ovarian cancer
The drug was approved by the USFDA in 2026 for the treatment of platinum-resistant ovarian cancer.[3]
Relacorilant is an orally available antagonist of the glucocorticoid receptor (GR), with potential antineoplastic activity. Upon administration, relacorilant competitively binds to and blocks GRs. This inhibits the activity of GRs, and prevents both the translocation of the ligand-GR complexes to the nucleus and gene expression of GR-associated genes. This decreases the negative effects that result from excess levels of endogenous glucocorticoids, like those seen when tumors overproduce glucocorticoids. In addition, by binding to GRs and preventing their activity, inhibition with CORT125134 also inhibits the proliferation of GR-overexpressing cancer cells. GRs are overexpressed in certain tumor cell types and promote tumor cell proliferation.
OriginatorCorcept Therapeutics
DeveloperCorcept Therapeutics; University of Chicago
Mechanism of ActionGlucocorticoid receptor antagonists
Orphan Drug StatusYes – Pancreatic cancer; Ovarian cancer; Cushing syndrome
RegisteredFallopian tube cancer; Ovarian cancer; Peritoneal cancer
PreregistrationCushing syndrome
Phase IIIAdenocarcinoma
Phase IIProstate cancer
DiscontinuedAdrenocortical carcinoma
27 Mar 2026Discontinued – Phase-I for Adrenocortical carcinoma (Inoperable/Unresectable, Late-stage disease, Metastatic disease, Combination therapy) in USA (PO), before March 2026 (Corcept Therapeutics pipeline, March 2026)
27 Mar 2026Corcept Therapeutics plans the phase II STELLA trial for Cervical cancer (Combination therapy, Second-line therapy or greater) in first quarter of 2026
25 Mar 2026Registered for Fallopian tube cancer (Combination therapy, Second-line therapy or greater) in USA (PO) – First global approval
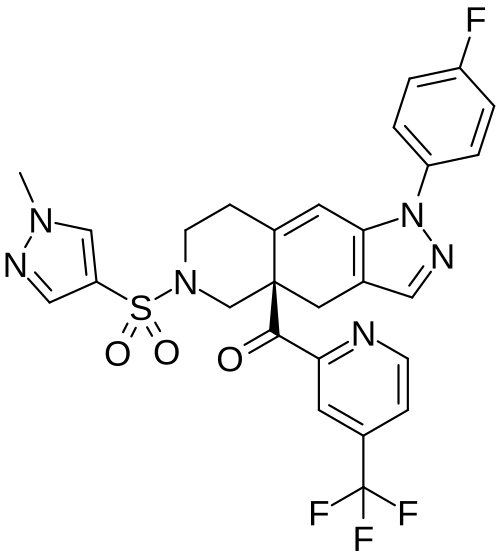
Relacorilant (CORT125134)118) is being developed by Corcept Therapeutics, Inc. It is an orally active, high-affinity, selective antagonist of the glucocorticoid receptor that may benefit from the modulation of cortisol activity. In structural optimization, the introduction of a trifluoromethyl group to the 4-position on the pyridyl moiety was found to increase HepG2 tyrosine amino transferase assay potency by a factor of four. Relacorilant is currently being evaluated in a phase II clinical study in patients with Cushing’s syndrome.119)
2-Bromo-4-(trifluoromethyl)pyridine (17) prepared from (E)-4-ethoxy-1,1,1-trifluorobut-3-en-2-one is employed as a key intermediate for the preparation of relacorilant as shown in Scheme 31.120)
118) H. Hunt, T. Johnson, N. Ray and I. Walters (Corcept Therapeutics, Inc.): PCT Int. Appl. WO2013/177559 (2013).
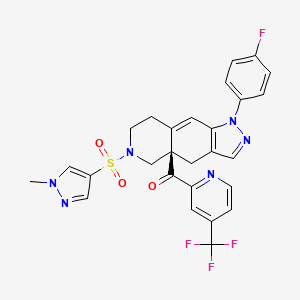
119) H. J. Hunt, J. K. Belanoff, I. Walters, B. Gourdet, J. Thomas, N. Barton, J. Unitt, T. Phillips, D. Swift and E. Eaton: Identification of the Clinical Candidate (R)-(1-(4-Fluorophenyl)-6-((1-methyl-1H-pyrazol-4-yl)sulfonyl)-4,4a,5,6,7,8-hexahydro-1H-pyrazolo[3,4-g]isoquinolin-4a-yl)(4-(trifluoromethyl)pyridin-2-yl)methanone (CORT125134): A Selective Glucocorticoid Receptor (GR) Antagonist. J. Med. Chem. 60, 3405–3421 (2017). [Abstract] [Google Scholar]
120) B. Lehnemann, J. Jung and A. Meudt (Archimica GmbH): PCT Int. Appl. WO 2007/000249 (2007).
The nonselective glucocorticoid receptor (GR) antagonist mifepristone has been approved in the U.S. for the treatment of selected patients with Cushing’s syndrome. While this drug is highly effective, lack of selectivity for GR leads to unwanted side effects in some patients. Optimization of the previously described fused azadecalin series of selective GR antagonists led to the identification of CORT125134, which is currently being evaluated in a phase 2 clinical study in patients with Cushing’s syndrome.
Cushing’s syndrome (CS) is a metabolic disorder caused by chronic hypercortisolism. CS is associated with cardiovascular, metabolic, skeletal and psychological dysfunctions and can be fatal if left untreated. The first-line treatment for all forms of CS is a surgery. However, medical therapy has to be chosen if surgical resection is not an option or is deemed ineffective. Currently available therapeutics are either not selective and have side effects or are only available as an injection (pasireotide).
 DRUG APPROVALS BY DR ANTHONY MELVIN CRASTO
.....FOR BLOG HOME CLICK HERE


 Subscribe in a reader






 This article incorporates text from this source, which is in the public domain.