New Drug Approvals

Home » Uncategorized (Page 86)

Category Archives: Uncategorized

DRUG APPROVALS BY DR ANTHONY MELVIN CRASTO .....FOR BLOG HOME CLICK HERE

Blog Stats

  • 5,049,701 hits

Flag and hits

Flag Counter

Enter your email address to follow this blog and receive notifications of new posts by email.

Join 37.7K other subscribers
Follow New Drug Approvals on WordPress.com

Archives

Categories

Recent Posts

Flag Counter

ORGANIC SPECTROSCOPY

Read all about Organic Spectroscopy on ORGANIC SPECTROSCOPY INTERNATIONAL 

Enter your email address to follow this blog and receive notifications of new posts by email.

Join 37.7K other subscribers
DR ANTHONY MELVIN CRASTO Ph.D

DR ANTHONY MELVIN CRASTO Ph.D

DR ANTHONY MELVIN CRASTO, Born in Mumbai in 1964 and graduated from Mumbai University, Completed his Ph.D from ICT, 1991,Matunga, Mumbai, India, in Organic Chemistry, The thesis topic was Synthesis of Novel Pyrethroid Analogues, Currently he is working with AFRICURE PHARMA, ROW2TECH, NIPER-G, Department of Pharmaceuticals, Ministry of Chemicals and Fertilizers, Govt. of India as ADVISOR, earlier assignment was with GLENMARK LIFE SCIENCES LTD, as CONSUlTANT, Retired from GLENMARK in Jan2022 Research Centre as Principal Scientist, Process Research (bulk actives) at Mahape, Navi Mumbai, India. Total Industry exp 32 plus yrs, Prior to joining Glenmark, he has worked with major multinationals like Hoechst Marion Roussel, now Sanofi, Searle India Ltd, now RPG lifesciences, etc. He has worked with notable scientists like Dr K Nagarajan, Dr Ralph Stapel, Prof S Seshadri, etc, He did custom synthesis for major multinationals in his career like BASF, Novartis, Sanofi, etc., He has worked in Discovery, Natural products, Bulk drugs, Generics, Intermediates, Fine chemicals, Neutraceuticals, GMP, Scaleups, etc, he is now helping millions, has 9 million plus hits on Google on all Organic chemistry websites. His friends call him Open superstar worlddrugtracker. His New Drug Approvals, Green Chemistry International, All about drugs, Eurekamoments, Organic spectroscopy international, etc in organic chemistry are some most read blogs He has hands on experience in initiation and developing novel routes for drug molecules and implementation them on commercial scale over a 32 PLUS year tenure till date Feb 2023, Around 35 plus products in his career. He has good knowledge of IPM, GMP, Regulatory aspects, he has several International patents published worldwide . He has good proficiency in Technology transfer, Spectroscopy, Stereochemistry, Synthesis, Polymorphism etc., He suffered a paralytic stroke/ Acute Transverse mylitis in Dec 2007 and is 90 %Paralysed, He is bound to a wheelchair, this seems to have injected feul in him to help chemists all around the world, he is more active than before and is pushing boundaries, He has 100 million plus hits on Google, 2.5 lakh plus connections on all networking sites, 100 Lakh plus views on dozen plus blogs, 227 countries, 7 continents, He makes himself available to all, contact him on +91 9323115463, email amcrasto@gmail.com, Twitter, @amcrasto , He lives and will die for his family, 90% paralysis cannot kill his soul., Notably he has 38 lakh plus views on New Drug Approvals Blog in 227 countries......https://newdrugapprovals.wordpress.com/ , He appreciates the help he gets from one and all, Friends, Family, Glenmark, Readers, Wellwishers, Doctors, Drug authorities, His Contacts, Physiotherapist, etc He has total of 32 International and Indian awards

Verified Services

View Full Profile →

Archives

Categories

Flag Counter

Lixivaptan


Lixivaptan structure.svg

Lixivaptan

CRTX-080; VPA-985; WAY-VPA-985

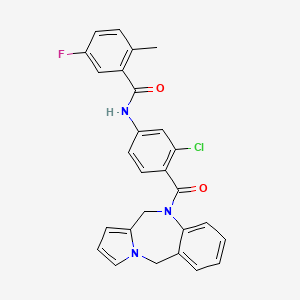
N-[3-chloro-4-(6,11-dihydropyrrolo[2,1-c][1,4]benzodiazepine-5-carbonyl)phenyl]-5-fluoro-2-methylbenzamide

 CAS 168079-32-1

MW 473.9,

 MF C27H21ClFN3O2

NDA Filing

A vasopressin (AVP) V2 antagonist potentially for treatment of heart failure and hyponatremia.

Lixivaptan (VPA-985) is a phase III pharmaceutical being developed by Cardiokine, Inc., a specialty pharmaceutical company based in Philadelphia, PA, focused on the development of pharmaceuticals for the treatment and prevention of cardiovascular diseases. Lixivaptan is, as of May 2010, in Phase III clinical trials involving patients with hyponatremia, including those with concomitant heart failure.[1] Hyponatremia is an electrolyte disturbance in which the sodium concentration in the serum is lower than normal. Lixivaptan may help some patients eliminate excess fluids while retaining electrolytes.

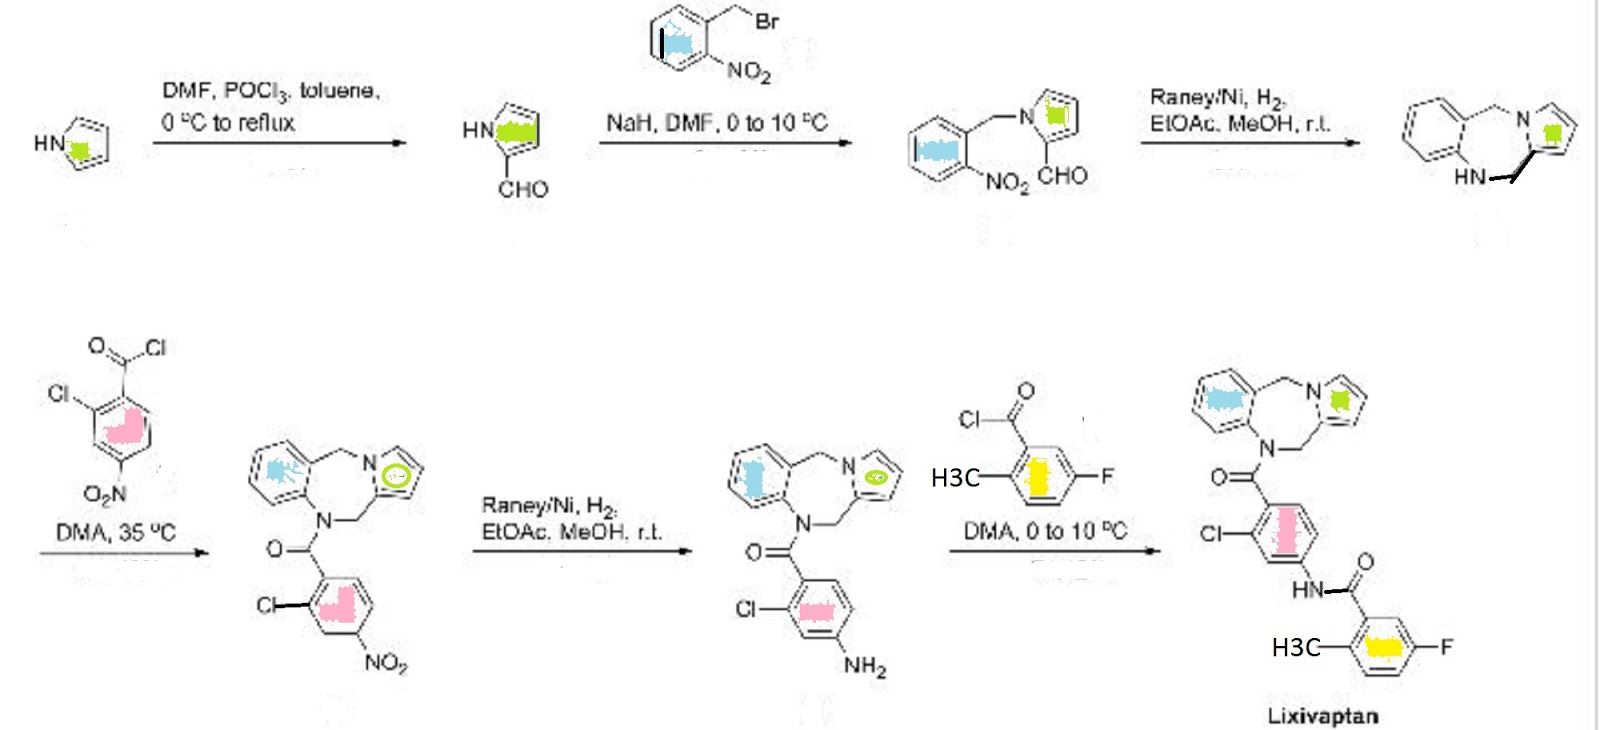
ChemistryLixivaptan is synthesized as follows:[2]

Lixivaptan rx.png


Mechanism of action

Lixivaptan is a potent, non-peptide, selective vasopressin 2 receptor antagonist. The oral capsule works by reducing the action of the hormone vasopressin that blocks fluid excretion. Lixivaptan acts by blocking vasopressin, an anti-diuretic hormone that causes the kidneys to retain water. When the body needs to remain hydrated under certain conditions, vasopressin can have protective effects. But an excess of vasopressin is counterproductive in a body retaining too much fluid. The drug shows promise in treating heart failure in patients with hyponatremia.

THE BALANCE study

In February 2008, Cardiokine and its worldwide partner, Biogen Idec, initiated THE BALANCE (Treatment of HyponatrEmia BAsed on LixivAptan in N Yha class III/IV Cardiac patient Evaluation) study. THE BALANCE study is a 650-patient Phase III, global, multi-center, randomized, placebo-controlled, double-blind, study of lixivaptan for hyponatremia in patients with heart failure. The primary objective is to evaluate the safety and effectiveness of lixivaptan, when compared to the placebo, in increasing serum sodium from baseline in heart failure patients with hyponatremia.[3][4]

Previous studies

In previous studies, lixivaptan improved blood sodium levels, lowered body weight and increased urine volume. Those studies suggest that lixivaptan may play an important role in treating hyponatremia and the signs and symptoms of water retention associated with heart failure, Syndrome of Inappropriate Anti-Diuretic Hormone(SIADH), and Liver Cirrhosis with Ascites (LCWA). In clinical trials involving patients with water volume overload, lixivaptan resulted in correction of hyponatremia together with marked aquaresis.

Vaptans

The vasopressin receptor antagonists, dubbed vaptans, target the vasopressin hormonal feedback system. Vasopressin, also called the anti-diuretic hormone or ADH, is an important part of regulation in the circulatory system and is integral to the balance of water in the body. As a fundamental part of hormonal control in the body, it is implicated in many different conditions. Vaptans can be administered orally or intravenously. They work by competing for the active sites on cells meant for vasopressin binding—in this way, the vasopressin is blocked from acting, which earns the title of vasopressing antagonists.

SYNTHESIS COMING………………..
JMC 1998, 41, 2442
US 5516774
CN103694240

Lixiputan (Lixivaptan, I) is pressurized by a Wyeth (wyeth) research and development of non-peptide hormone arginine oral selective V2 receptor antagonist, chemical name N- [3- chloro-4- (10, 11- dihydro -5H- pyrrolo [2,1-c] [1,4] benzodiazepine-10-yl carbonyl) phenyl] -5-fluoro-2- methylbenzamide. Clinical studies have shown that, compared with traditional diuretic, Lixiputan for the treatment of congestive heart failure (CHF), cirrhosis of hyponatremia and syndrome of inappropriate antidiuretic hormone secretion disorders (SIADH) patients, its in increase free water clearance without affecting renal sodium discharge, it will not activate the neuroendocrine system, and has a high safety and tolerability. Lixiputan V2 receptor selectivity higher than in May 2009 the FDA approved tolvaptan, Phase III clinical studies for the treatment of hyponatremia have been completed in the United States, in the pre-registration stage.

Document (Journalof medicinal chemistry, 1998,41 (14):. 2442-2444) reported Lixiputan there are two synthetic routes, one route to 10,11-dihydro -5H- pyrrolo [2, ι-c] [1,4] benzodiazepine (2) as raw materials, in turn with 2-chloro-4-nitrobenzoyl and 5-fluoro-2-methylbenzoyl docking, to obtain I; the second is the first line of 2-chloro-4-amino benzoic acid methyl ester (5) and 5-fluoro-2-methylbenzoyl chloride (7) butt, by hydrolysis, acylation reaction of 2-chloro-like -4 – [(5-fluoro-2-methylbenzoyl) amino] benzoyl chloride (10), and then with 2 reaction of I. 2 As the raw material is expensive, Route One to two as the starting material, the multi-step reaction, its low efficiency, high cost of production. Therefore, this study reference line two, 2-chloro-4-nitro-benzoic acid (3) as the starting material, by esterification, hydrogenation, acylation, hydrolysis, chloride, and so the reaction of 10; 10 and then with 2 After acylation reaction of N- I. I synthetic route follows.


Figure CN103694240AD00041

The chemical structure:


Figure CN103694240AD00042

formula = C27H21ClFN3O2

 Molecular Weight: 473.93

The method for producing foreign products have been reported, such as the literature Journal of medicinalchemistry, 1998,41 (14):. 2442-2444 and US, 5516774 [P], 1996-5-14. Currently, Lixiputan (Iixivaptan) abroad in Phase III clinical studies, there are good prospects for development, given the value of the pharmaceutical compounds, high purity, with a very determined and reproducible crystalline compounds are important .

The present inventors have repeated the document US, 5,516,774 Lixiputan method of purity, obtained was 97.5%, mpl91-195 ° C, by the study of a plurality of batches, the melting point of the same, by a powder X- ray diffraction pattern See

preparation of Lixiputan solvate Lixiputan, by two synthetic methods. As literature Journalof medicinal chemistry, 1998, 41 (14):. 2442-2444 and US, 5516774 [P],

The method reported in [0026] 1996-5-14. Preclude the use of the route of the present invention is represented by the following reaction:


Figure CN103694240AD00071

  synthetic Lixiputan by proton nuclear magnetic resonance spectroscopy (1H-NMRX mass spectrometry (MS), infrared spectroscopy (IR) and other confirmed its chemical structure (see Figure 3 MS). Test equipment for nuclear magnetic resonance Bruker AV400 meter, gas generation agent for CambridgeIsotope Laboratories Company DMS0_d6.

  ES1-HRMS (m / z): 474.17 [M + H] + NMR (400MHz, DMS0_d6) δ: 10.49 (s, 1H), 7.84 (s, 1H), 7.40 (d, J = 6.8Hz, 2H), 7.33 (d, J = 8.4Hz, 3H), 7.23 (t, J = 8.4Hz, 1H), 7.13 (t, J = 5.6Hz, 2H), 7.05 (d, J = 6.8Hz, 1H) , 6.82 (s, 1H), 5.94 (d, J = 32Hz, 2H), 5.23 (br, 4H), 2.30 (s, 3H).

The product obtained, with a purity of 97.5%, mp 191-195 ° C.

Figure CN103694240AD00072

Lixiputan solvates H NMR spectrum, δ: 1.147-1.182 “3” methyl hydrogens; δ: 1.971-1.977 for the “I” position methyl hydrogen; δ: 3.994-4.047 “2” position methylene hydrogen.

CN104059070
CN104140429
IN 2012 MUM 03309
Lixivaptan.png

References

Patent Submitted Granted
Tricyclic diazepine vasopressin antagonists and oxytocin antagonists [US5854237] 1998-12-29
Tricyclic diazepine vasopressin antagonists and oxytocin antagonists [US5889001] 1999-03-30
Tricyclic diazepine vasopressin antagonists and oxytocin antagonists [US5843944] 1998-12-01
Tricyclic diazepine vasopressin antagonists and oxytocin antagonists [US5624923] 1997-04-29
Compositions for delivery of insoluble agents [US8877746] 2010-08-24 2014-11-04
Patent Submitted Granted
AURIS FORMULATIONS FOR TREATING OTIC DISEASES AND CONDITIONS [US2009306225] 2009-12-10
Vasopressin antagonist and diuretic combination [US6656931] 2003-04-10 2003-12-02
Pharmaceutical carrier formulation [US6437006] 2002-08-20
Vasopressin antagonist formulation and process [US6352718] 2002-03-05
Nonpeptide agonists and antagonists of vasopressin receptors [US2002128208] 2002-09-12
Tricyclic diazepine vasopressin antagonists and oxytocin antagonists [US5968930] 1999-10-19
Tricyclic diazepine vasopressin antagonists and oxytocin antagonists [US5968937] 1999-10-19
Tricyclic diazepine vasopressin antagonists and oxytocin antagonists [US5516774] 1996-05-14
Tricyclic diazepine vasopressin antagonists and oxytocin antagonists [US5733905] 1998-03-31
Tricyclic diazepine vasopressin antagonists and oxytocin antagonists [US5736540] 1998-04-07
Lixivaptan
Lixivaptan structure.svg
Systematic (IUPAC) name
N-[3-chloro-4-(6,11-dihydropyrrolo[2,1-c][1,4]benzodiazepine-5-carbonyl)phenyl]-5-fluoro-2-methylbenzamide
Identifiers
CAS Number 168079-32-1 
ATC code None
PubChem CID: 172997
IUPHAR/BPS 2238
ChemSpider 151067 
UNII 8F5X4B082E Yes
ChEMBL CHEMBL49429 
Chemical data
Formula C27H21ClFN3O2
Molecular mass 473.926 g/mol
CN102020609A * Sep 17, 2009 Apr 20, 2011 北京本草天源药物研究院 Tolvapta crystal or amorphous substance and preparation method thereof
CN102918038A * Mar 31, 2011 Feb 6, 2013 万梯雅有限公司 New polymorph
US5516774 * Jun 13, 1994 May 14, 1996 American Cyanamid Company Tricyclic diazepine vasopressin antagonists and oxytocin antagonists
1 * 吕扬 等: “《晶型药物》”, 31 October 2009, article “”第七章 晶型药物的研究方法”“, pages: 136-139
//////////Lixivaptan, CRTX-080,  VPA-985,  WAY-VPA-985
CC1=C(C=C(C=C1)F)C(=O)NC2=CC(=C(C=C2)C(=O)N3CC4=CC=CN4CC5=CC=CC=C53)Cl

Uridine triacetate, ウリジントリアセタート FDA approves first emergency treatment for overdose of certain types of chemotherapy


Uridine triacetate.svg
12/11/2015 12:05 PM EST
The U.S. Food and Drug Administration today approved Vistogard (uridine triacetate) for the emergency treatment of adults and children who receive an overdose of the cancer treatment fluorouracil or capecitabine, or who develop certain severe or life-threatening toxicities within four days of receiving these cancer treatments.

December 11, 2015

Release

The U.S. Food and Drug Administration today approved Vistogard (uridine triacetate) for the emergency treatment of adults and children who receive an overdose of the cancer treatment fluorouracil or capecitabine, or who develop certain severe or life-threatening toxicities within four days of receiving these cancer treatments.

“Treating cancer requires not only selecting which drug may be most effective and well tolerated, but ensuring the correct dose is given at proper intervals. While rare, unintentional overdose can occur,” said Richard Pazdur, M.D., director of the Office of Hematology and Oncology Products in the FDA’s Center for Drug Evaluation and Research. “Today’s approval is a first-of-its-kind therapy that can potentially save lives following overdose or life-threatening toxicity from these chemotherapy agents.”

Fluorouracil (taken by infusion) and capecitabine (taken orally) are similar types of chemotherapy that have been used for decades to treat several types of cancer, including breast and gastrointestinal cancers. An overdose of fluorouracil or capecitabine is rare, but when it occurs, the effects are serious and can be fatal.

Vistogard, taken orally, blocks cell damage and cell death caused by fluorouracil chemotherapy. Patients should take Vistogard as soon as possible after the overdose (whether or not they have symptoms) or early-onset (within four days) of severe or life-threatening toxicity. The patient’s health care provider will determine when he or she should return to the prescribed chemotherapy after treatment with Vistogard.

The efficacy and safety of Vistogard were studied in 135 adult and pediatric cancer patients who were treated in two separate trials and had either received an overdose of flourouracil or capecitabine, or had early-onset, unusually severe or life-threatening toxicities within 96 hours after receiving flourouracil (not due to an overdose). The studies’ primary measure was survival at 30 days or until chemotherapy could resume if prior to 30 days. Of those who were treated with Vistogard for overdose, 97 percent were still alive at 30 days. Of those treated with Vistogard for early-onset severe or life-threatening toxicity, 89 percent were alive at 30 days. In both studies, 33 percent of patients resumed chemotherapy in less than 30 days.

Vistogard is not recommended for treating non-emergency adverse reactions associated with flourouracil or capecitabine because Vistogard may lessen the efficacy of these drugs. The safety and efficacy of Vistogard initiated more than 96 hours following the end of treatment with flourouracil or capecitabine have not been established.

The most common side effects of treatment with Vistogard were diarrhea, vomiting and nausea.

The FDA granted Vistogard orphan drug designation, which provides financial incentives, like clinical trial tax credits, user fee waivers, and eligibility for market exclusivity to promote rare disease drug development. Vistogard was also granted priority review and fast track designations, which are distinct programs intended to facilitate and expedite the development and review of certain new drugs in light of their potential to benefit patients with serious or life-threatening conditions.

Vistogard is marketed by Wellstat Therapeutics Corporation based in Gaithersburg, Maryland.

 UPDATED IN SEPT 2016…………..
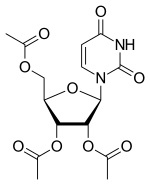
 ChemSpider 2D Image | uridine triacetate | C15H18N2O9
2',3',5'-Tri-O-acetyluridine.png
Uridine triacetate
Uridine, 5-hydroxy-, 2′,3′,5′-triacetate
2′,3′,5′-Tri-O-acétyluridine
223-881-5 [EINECS]
CAS 4105-38-8
Priority review drug 
Orphan drug
FAST TRACK
MF C15H18N2O9, MW 370.314
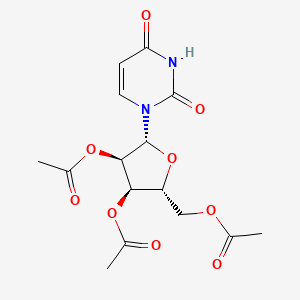
[(2R,3R,4R,5R)-3,4-bis(acetyloxy)-5-(2,4-dioxo-1,2,3,4-tetrahydropyrimidin-1-yl)oxolan-2-yl]methyl acetate
Vistogard [Trade name]
Xuriden [Trade name]
(2R,3R,4R,5R)-2-(acetoxymethyl)-5-(2,4-dioxo-3,4-dihydropyrimidin-1(2H)-yl)tetrahydrofuran-3,4-diyl diacetate
Wellstat (Originator)
PN-401; RG-2133; TAU
MOA:Pyrimidine analog
Indication:Hereditary orotic aciduria; Chemotherapy induced poisoning
To treat patients with hereditary orotic aciduria
Drug Name(s) XURIDEN
FDA Application No. (NDA) 208169
Active Ingredient(s) URIDINE TRIACETATE
Company WELLSTAT THERAP
Original Approval or Tentative Approval Date September 4, 2015

FDA APPROVAL SUMMARY

Chemotherapy induced poisoning, VISTOGARD, FDA 2015-12-11

Hereditary orotic aciduria, Xuriden, FIRST APPROVAL, 2015-09-04

 Image result for SYNTHESIS, Uridine triacetate

2′,3′,5′-tri-O-acetyluridine
2′,3′,5′-Triacetyluridine
Tri-O-acetyluridine
Triacetyl uridine
Triacetyluridine
Uridine 2′,3′,5′-triacetate
Vistonuridine
External Identifiers
  • PN 401
  • PN-401
  • PN401
  • RG 2133
  • RG-2133
  • RG2133

Uridine triacetate is a drug used in the treatment of hereditary orotic aciduria[1] and to treat patients following an overdose ofchemotherapy drugs 5-fluorouracil or capecitabine, or in patients exhibiting early-onset, severe or life-threatening toxicity affecting the cardiac or central nervous system, and/or early-onset, unusually severe adverse reactions (e.g., gastrointestinal toxicity and/or neutropenia) within 96 hours following the end of 5-fluorouracil or capecitabine administration.[2][3]

Uridine triacetate was developed, manufactured and distributed by Wellstat Therapeutics and it is marketed in USA by BTG. Also, It was granted breakthrough therapy designation by FDA in 2015.

Uridine triacetate is a prodrug of uridine.[4]

Uridine triacetate, formerly known as vistonuridine, is an orally active prodrug of the naturally occurring nucleoside uridine. It is used for the treatment of hereditary orotic aciduria (Xuriden), or for the emergency treatment of fluorouracil or capecitabine overdose or toxicity (Vistogard). It is provided in the prodrug form as uridine triacetate as this form delivers 4- to 6-fold more uridine into the systemic circulation compared to equimolar doses of uridine itself. When used for the treatment or prevention of toxicity associated with fluorouracil and other antimetabolites, uridine triacetate is utilized for its ability to compete with 5-fluorouracil (5-FU) metabolites for incorporation into the genetic material of non-cancerous cells. It reduces toxicity and cell-death associated with two cytotoxic intermediates: 5-fluoro-2′-deoxyuridine-5′-monophosphate (FdUMP) and 5-fluorouridine triphosphate (FUTP). Normally, FdUMP inhibits thymidylate synthase required for thymidine synthesis and DNA replication and repair while FUTP incorporates into RNA resulting in defective strands. As a result, these metabolites are associated with various unpleasant side effects such as neutropenia, mucositis, diarrhea, and hand–foot syndrome. Like many other neoplastic agents, these side effects limit the doses of 5-FU that can be administered, which also affects the efficacy for treatment. By pre-administering with uridine (as the prodrug uridine triacetate), higher doses of 5-FU can be given allowing for improved efficacy and a reduction in toxic side effects [3]. It can also be used as a rescue therapy if severe side effects present within 96 hours after initiation of therapy. Uridine triacetate is also used for the treatment of hereditary orotic aciduria, also known as uridine monophosphate synthase deficiency. This rare congenital autosomal recessive disorder of pyrimidine metabolism is caused by a defect in uridine monophosphate synthase (UMPS), a bifunctional enzyme that catalyzes the final two steps of the de novo pyrimidine biosynthetic pathway. As a result of UMPS deficiency, patients experience a systemic deficiency of pyrimidine nucleotides, accounting for most symptoms of the disease. Additionally, orotic acid from the de novo pyrimidine pathway that cannot be converted to UMP is excreted in the urine, accounting for the common name of the disorder, orotic aciduria. Furthermore, orotic acid crystals in the urine can cause episodes of obstructive uropathy. When administered as the prodrug uridine triacetate, uridine can be used by essentially all cells to make uridine nucleotides, which compensates for the genetic deficiency in synthesis in patients with hereditary orotic aciduria. When intracellular uridine nucleotides are restored into the normal range, overproduction of orotic acid is reduced by feedback inhibition, so that urinary excretion of orotic acid is also reduced.

Image result for SYNTHESIS, Uridine triacetate

Marketed as the product Xuriden (FDA), uridine triacetate is indicated for the treatment of hereditary orotic aciduria. Marketed as the product Vistogard (FDA), uridine triacetate is indicated for the emergency treatment of adult and pediatric patients in the following situations: following a fluorouracil or capecitabine overdose regardless of the presence of symptoms; or who exhibit early-onset, severe or life-threatening toxicity affecting the cardiac or central nervous system, and/or early-onset, unusually severe adverse reactions (e.g., gastrointestinal toxicity and/or neutropenia) within 96 hours following the end of fluorouracil or capecitabine administration.

Image result for SYNTHESIS, Uridine triacetateImage result for SYNTHESIS, Uridine triacetate

Uridine Triacetate was approved by the U.S. Food and Drug Administration (FDA) on Sep 4, 2015. It was developed by Wellstat Therapeutics, then marketed as Xuriden® by Wellstat Therapeutics in US. Then it was also approved by FDA for overdose of certain types of chemotherapy on Dec 11, 2015 and marketed as Vistogard®.

Uridine Triacetate is a prodrug of the nucleoside uridine used to treat hereditary orotic aciduria. Hereditary orotic aciduria is inherited from a recessive gene. The disease is due to a defective or deficient enzyme, which results in the body being unable to normally synthesize uridine, a necessary component of ribonucleic acid (RNA). Signs and symptoms of the disease include blood abnormalities (anemia, decreased white blood cell count, decreased neutrophil count), urinary tract obstruction due to the formation of orotic acid crystals in the urinary tract, failure to thrive, and developmental delays.

Xuriden® is approved as oral granules that can be mixed with food or in milk or infant formula, and is administered once daily. The starting dosage is 60 mg/kg once daily; the dose may be increased to 120 mg/kg (not to exceed 8 grams) once daily for insufficient efficacy.

Mechanism Of Action

Uridine triacetate is an acetylated form of uridine. Following oral administration, uridine triacetate is deacetylated by nonspecific esterases present throughout the body, yielding uridine in the circulation (Figure 1).

Figure 1: Uridine Triacetate Conversion to Uridine

Uridine Triacetate Conversion to Uridine - Illustration

URIDEN provides uridine in the systemic circulation of patients with hereditary orotic aciduria who cannot synthesize adequate quantities of uridine due to a genetic defect in uridine nucleotide synthesis.

Uridine triacetate is a synthetic uridine pro-drug that is converted to uridine in vivo. When used for the treatment or prevention of toxicity associated with fluorouracil and other antimetabolites, uridine triacetate is utilized for its ability to compete with 5-fluorouracil (5-FU) metabolites for incorporation into the genetic material of non-cancerous cells. It reduces toxicity and cell-death associated with two cytotoxic intermediates: 5-fluoro-2′-deoxyuridine-5′-monophosphate (FdUMP) and 5-fluorouridine triphosphate (FUTP). By pre-administering with uridine (as the prodrug uridine triacetate), higher doses of 5-FU can be given allowing for improved efficacy and a reduction in toxic side effects [A18578] such as neutropenia, mucositis, diarrhea, and hand–foot syndrome. Uridine triacetate is also used for replacement therapy in the treatment of hereditary orotic aciduria, also known as uridine monophosphate synthase (UMPS) deficiency. As a result of UMPS deficiency, patients experience a systemic deficiency of pyrimidine nucleotides, accounting for most symptoms of the disease. Additionally, orotic acid from the de novo pyrimidine pathway that cannot be converted to UMP is excreted in the urine, accounting for the common name of the disorder, orotic aciduria. Furthermore, orotic acid crystals in the urine can cause episodes of obstructive uropathy. When administered as the prodrug uridine triacetate, uridine can be used by essentially all cells to make uridine nucleotides, which compensates for the genetic deficiency in synthesis in patients with hereditary orotic aciduria.

Route 1

Reference:1. J. Am. Chem. Soc. 1953, 75, 2017-2019.

2. Angew. Chem. internat. Edit. 1971, 10, 75.

3. US3116282.

PATENT

Production Example 1

Figure US06900298-20050531-C00001

5.6 g of uracil and 0.1 g of ammonium sulfate were dissolved in 22.4 ml of 1,1,1,3,3,3-hexamethyldisilazane and reacted at 120° C. for 2.5 hours. After the completion of the reaction, the reaction mixture was distilled to give 11.8 g of 2,4-bis(trimethylsilyloxy)-1,3-diazine. 1H-NMR (400 MHz, in C2D6CO): δ=0.29 (s, 9H), 0.31 (s, 9H), 6.35 (d, J=5.6 Hz, 1H), 8.19 (d, J=5.5Hz, 1H)

Referential Example 11.21 g of 2,4-bis(trimethylsilyloxy)-1,3-diazine obtained in PRODUCTION EXAMPLE 1 and 1.15 g of 1,2,3,5-tetra-O-acetyl-β-D-ribofuranose were dissolved in 4.8 ml of acetonitrile and cooled to 5° C. Next, 0.94 g of SnCl4 was added dropwise thereinto at the same temperature. After stirring for 10 minutes at the same temperature, the mixture was heated to 50° C. and reacted for 3 hours. The reaction mixture was analyzed by HPLC. Thus, β-uridine triacetate was obtained with a reaction yield of 83%.

Example 1

Figure US06900298-20050531-C00002

0.93 g of 2,4-bis(trimethylsilyloxy)-1,3-diazine obtained in PRODUCTION EXAMPLE 1 and 0.92 g of 1,2,3,5-tetra-O-acetyl-β-D-ribofuranose were dissolved in 4.7 ml of acetonitrile and cooled to 4° C. Then 0.49 g of FeCl3 was added thereto at the same temperature. After stirring for 10 minutes at the same temperature, the mixture was heated to 50° C. and reacted. The reaction was monitored by HPLC. After the completion of the reaction, the reaction mixture was added dropwise at 4° C. into a cold aqueous solution of sodium hydrogencarbonate which had been preliminarily prepared. After filtering off the catalyst residue, the filtrate was separated and the aqueous layer was extracted with 20 ml portions of ethyl acetate thrice. The organic layers were combined, washed with a saturated aqueous solution of sodium chloride and dried over sodium sulfate. After distilling off the solvent, 1.2 g (purity 80%) of the target compound was obtained as a viscous white solid.

Namely, the target compound could be obtained at a yield comparable to REFERNTIAL EXAMPLE 1 wherein SnCl4 was employed as the catalyst. 1H-NMR (400 MHz, in CDCl3): δ=2.11 (s, 3H), 2.14 (s, 3H), 2.15 (s, 3H), 4.35 (m, 3H), 5.33 (m, 2H), 5.79 (d, J=8.2 Hz, 1H), 6.04 (d, J=4.9 Hz, 1H), 7.39 (d, J=8.2 Hz, 1H)

Image result for SYNTHESIS, Uridine triacetate

CLIP

12/11/2015 12:05 PM EST
The U.S. Food and Drug Administration today approved Vistogard (uridine triacetate) for the emergency treatment of adults and children who receive an overdose of the cancer treatment fluorouracil or capecitabine, or who develop certain severe or life-threatening toxicities within four days of receiving these cancer treatments.

December 11, 2015

Release

The U.S. Food and Drug Administration today approved Vistogard (uridine triacetate) for the emergency treatment of adults and children who receive an overdose of the cancer treatment fluorouracil or capecitabine, or who develop certain severe or life-threatening toxicities within four days of receiving these cancer treatments.

“Treating cancer requires not only selecting which drug may be most effective and well tolerated, but ensuring the correct dose is given at proper intervals. While rare, unintentional overdose can occur,” said Richard Pazdur, M.D., director of the Office of Hematology and Oncology Products in the FDA’s Center for Drug Evaluation and Research. “Today’s approval is a first-of-its-kind therapy that can potentially save lives following overdose or life-threatening toxicity from these chemotherapy agents.”

Fluorouracil (taken by infusion) and capecitabine (taken orally) are similar types of chemotherapy that have been used for decades to treat several types of cancer, including breast and gastrointestinal cancers. An overdose of fluorouracil or capecitabine is rare, but when it occurs, the effects are serious and can be fatal.

Vistogard, taken orally, blocks cell damage and cell death caused by fluorouracil chemotherapy. Patients should take Vistogard as soon as possible after the overdose (whether or not they have symptoms) or early-onset (within four days) of severe or life-threatening toxicity. The patient’s health care provider will determine when he or she should return to the prescribed chemotherapy after treatment with Vistogard.

The efficacy and safety of Vistogard were studied in 135 adult and pediatric cancer patients who were treated in two separate trials and had either received an overdose of flourouracil or capecitabine, or had early-onset, unusually severe or life-threatening toxicities within 96 hours after receiving flourouracil (not due to an overdose). The studies’ primary measure was survival at 30 days or until chemotherapy could resume if prior to 30 days. Of those who were treated with Vistogard for overdose, 97 percent were still alive at 30 days. Of those treated with Vistogard for early-onset severe or life-threatening toxicity, 89 percent were alive at 30 days. In both studies, 33 percent of patients resumed chemotherapy in less than 30 days.

Vistogard is not recommended for treating non-emergency adverse reactions associated with flourouracil or capecitabine because Vistogard may lessen the efficacy of these drugs. The safety and efficacy of Vistogard initiated more than 96 hours following the end of treatment with flourouracil or capecitabine have not been established.

The most common side effects of treatment with Vistogard were diarrhea, vomiting and nausea.

The FDA granted Vistogard orphan drug designation, which provides financial incentives, like clinical trial tax credits, user fee waivers, and eligibility for market exclusivity to promote rare disease drug development. Vistogard was also granted priority review and fast track designations, which are distinct programs intended to facilitate and expedite the development and review of certain new drugs in light of their potential to benefit patients with serious or life-threatening conditions.

Vistogard is marketed by Wellstat Therapeutics Corporation based in Gaithersburg, Maryland.

CLIP

With support from Almac, Wellstat delivers for a rare disease.

Proximity of API and finished drug development helps uridine triacetate to market for two indications

By Rick Mullin

“The initial contact was a cold call by Almac in 2010 or 2011,” recalls Mike Bamat, senior vice president of R&D at Wellstat Therapeutics, a small drug company in Gaithersburg, Md. “There were probably a couple of calls. It was one of those things where timing is everything.”

Almac, a Craigavon, Northern Ireland-based pharmaceutical services company, was looking to get in on Wellstat’s development of uridine triacetate, a synthetic pyrimidine analog, as an antidote for fluorouracil and capecitabine toxicity and overdose in cancer patients receiving those chemotherapies. And the calls, which Almac records indicate followed some communication between the companies, happened to come just when Wellstat was looking to change service partners as it moved toward commercial development of the drug.


Uridine triacetate

Discovery: Wellstat Therapeutic’s research on the therapeutic potential of exogenous uridine leads to a determination that uridine triacetate is a safe means of delivering the agent
Applications: Treatment of hereditary orotic aciduria (HOA), an extremely rare disease in which the body does not produce uridine, causing overproduction of orotic acid; emergency treatment of toxic reaction to or overdose of the cancer treatments fluorouracil and capecitabine
Methods of action: Treating HOA, uridine triacetate restores intracellular nucleotide concentrations, normalizing orotic acid production; as a chemotherapy antidote, it increases intracellular levels of uridine to dilute fluorouracil and capecitabine
Years in development: Since 2008 for chemotherapy antidote, and 2013 for HOA
Approved: Xuriden for HOA, Sept. 4, 2015; Vistogard for chemotherapy antidote, Dec. 11, 2015


The job went to Almac, as did work that sprang up as the result of another phone call to Wellstat—this one from the U.S. Food & Drug Administration.

As Bamat explains, uridine triacetate caught FDA’s attention regarding another potential indication—an extremely rare and life-threatening disease called hereditary orotic aciduria, or HOA. A consequence of the body’s inability to produce uridine, a necessary component of ribonucleic acid, HOA can manifest in a range of symptoms including blood abnormalities, developmental delays, and urinary tract obstruction caused by overproduction of orotic acid. There have been 20 reported cases of HOA since the 1950s. Only four cases are currently known in the U.S., Bamat says, and likely fewer than 20 in the world.

Wellstat landed approvals for Xuriden, the HOA treatment, in September of last year and Vistogard, the chemotherapy antidote, in December.

The story of Xuriden centers on a raft of FDA incentives for super-rare diseases that enabled Wellstat to move forward on an expedited application for a drug that will never be made in any great volume. But bringing Xuriden and Vistogard to market may also be viewed as the story of a drug discovery firm becoming a commercial enterprise thanks to its partnership with a service provider.

As Wellstat began late-stage development of the chemotherapy antidote, its research partner at the time, QS Pharma, was acquired by the service firm WIL Research. The look and feel of the partnership changed, according to Bamat.

“We kind of lost the small, easy-to-work-with relationship we had with them,” he says. Wellstat also needed support on development and manufacturing of a finished drug product composed of granules delivered in packets or sachets. The drug is administered orally, usually sprinkled on food such as applesauce or yogurt.

Almac was deemed a good fit because of its experience with developing drugs in granule form for “sachet presentation,” a packaging method more common in Europe than in the U.S. The Northern Ireland firm’s ability to develop and manufacture the active pharmaceutical ingredient (API) and the drug product in one location—at its headquarters—would also prove to be a significant advantage.

The distance between Gaithersburg and Craigavon, however, was a concern, according to Bamat. “We debated it. Especially those of us who knew we would be going there,” he says. “We couldn’t just jump in a car and go. But we looked at a variety of things, including cost and value, and it was all very positive at Almac.”

According to David Downey, vice president of commercial operations at Almac, bringing Wellstat’s work on uridine triacetate to commercial production posed several challenges, the first being to secure supply of uridine starting material, which is extracted from sugar beets by Euticals, an Italian firm. Next was developing a method to control particle size in both the API and the finished product. Almac also had to validate process equipment as it scaled up production.

“Uridine triacetate is Wellstat’s first commercial product,” Downey says. “So we were provided with a process more fit for development than for commercial production.”

The basic formulation of a granule drug product is simple, according to Downey: The API and excipient are mixed in a dry blender. The challenge is developing an analytical regimen to assure the granules are blended uniformly. Meeting the challenge required a high level of coordination between API and drug product process development.

“Wellstat needed a partner that could support them from the API to the drug product,” Downey says. The physical proximity between the Almac facilities in Craigavon conducting API and drug product work was a key advantage, he claims.

09414-cover-drugscxd
Uridine triacetate is formulated into granules presented in packets and sprinkled on food.
Credit: Wellstat Therapeutics

“If you listen to our business development people, you’ll hear them use the term, ‘crossing car parks as opposed to crossing oceans,’ ” Downey says, explaining that many competitors who offer API and finished drug services run these operations thousands of kilometers apart from each other, sometimes on different continents.

Before it signed on with Almac, Wellstat had been working with uridine triacetate for about 10 years. Its focus on developing the antidote drug started in 2008. Branching into the HOA treatment, however, upped the stakes.

Clinical study development for an HOA therapy was expedited via a full house of regulatory incentives from FDA, according to Bamat. “We had orphan drug designation, rare pediatric designation, breakthrough therapy designation, and priority review,” he says. “So they really went all out in helping us develop this.”

Although Wellstat was interested in developing a life saving drug for children, it was concerned about paying for it, given the tiny market. “At that time, the rare pediatric disease priority review voucher program was just on the radar,” Bamat says. “FDA said, ‘Consider this new program. Maybe it’s a way that at some risk you could recoup some of your costs.’ We looked at it and were willing to take the risk.”

It paid off. Wellstat was able to sell its priority review voucher—which entitles a company that brings a rare pediatric drug to market to receive expedited review of a subsequent drug—to AstraZeneca last year for an undisclosed amount. Other vouchers sold in 2015 brought high sums, including $350 million for one that AbbVie bought from United Therapeutics in August.

Bamat says Wellstat is not likely to change focus after its success with uridine triacetate. It continues to investigate new indications for the compound and will likely work with Almac on anything going into commercial development.

He emphasizes the importance of maintaining an effective working relationship with an outsourcing partner. “My main consideration is that these are people we can really work with on a day-to-day, week-to-week basis,” Bamat says. “Will the communication be good? Will they be honest and transparent with us, and will we be the same for them? That was a key factor, and we felt it was a plus with Almac.”

Uridine triacetate
Uridine triacetate.svg
Clinical data
Trade names Vistogard, Xuriden
Routes of
administration
Oral granules
Legal status
Legal status
Pharmacokinetic data
Metabolism Pyrimidine catabolic pathway
Onset of action Tmax = 2-3 hours
Biological half-life 2-2.5 hours
Excretion Renal
Identifiers
DrugBank DB09144
Chemical data
Formula C15H18Cl0N2O9S0
Molar mass 370.31 g·mol−1

References

  1.  HIGHLIGHTS OF PRESCRIBING INFORMATION OF XURIDEN
  2. Jump up^ BTG Announces FDA Approval of VISTOGARD® (Uridine Triacetate) as Antidote to Overdose and Early Onset, Severe, or Life-Threatening Toxicities from Chemotherapy Drugs 5-Fluorouracil (5-FU) or Capecitabine
  3. Jump up^ “FDA Approved Drugs:Uridine Triacetate”. FDA. 2015-12-11. Retrieved 2016-04-29.
  4.  “Uridine triacetate”. DrugBank.
Patent ID Date Patent Title
EP0339075 1993-08-18 ACYLATED URIDINE AND CYTIDINE AND USES THEREOF
US5200514 1993-04-06 SYNTHESIS OF 2′-DEOXYPYRIMIDINE NUCLEOSIDES
US5047520 1991-09-10 2′-alkylidenepyrimidine nucleoside derivatives, process for production thereof, and uses thereof
EP0204264 1990-08-16 CONVERSION OF URACIL DERIVATIVES TO CYTOSINE DERIVATIVES
WO8903837 1989-05-05 ACYLATED URIDINE AND CYTIDINE AND USES THEREOF
US4754026 1988-06-28 Conversion of uracil derivatives to cytosine derivatives
Patent ID Date Patent Title
US7807654 2010-10-05 Compositions and methods for treatment of mitochondrial diseases
US2010222296 2010-09-02 PYRIMIDINES, SUCH AS URIDINE, IN TREATMENTS FOR PATIENTS WITH BIPOLAR DISORDER
US7737128 2010-06-15 Pyrimidines, such as uridine, in treatments for patients with bipolar disorder
US2010098678 2010-04-22 Methods of Treatment of Mitochondrial Disorders
US2010041620 2010-02-18 METHODS FOR IMPROVING FRONTAL BRAIN BIOENERGETIC METABOLISM
US2010041621 2010-02-18 METHODS AND COMPOSITIONS FOR IMPROVING COGNITIVE PERFORMANCE
US7582619 2009-09-01 Compositions and methods for treatment of mitochondrial diseases
US2008226684 2008-09-18 METHOD AND PROCESS FOR THE PRODUCTION OF MULTI-COATED RECOGNITIVE AND RELEASING SYSTEMS
US7105498 2006-09-12 Acylated uridine and cytidine and uses thereof
US6956028 2005-10-18 Compositions and methods for treatment of mitochondrial diseases
Patent ID Date Patent Title
US2015307542 2015-10-29 MODIFIED NUCLEIC ACID MOLECULES AND USES THEREOF
US2015167017 2015-06-18 ALTERNATIVE NUCLEIC ACID MOLECULES AND USES THEREOF
US8821899 2014-09-02 Method and process for the production of multi-coated recognitive and releasing systems
US8771713 2014-07-08 Method and process for the production of multi-coated recognitive and releasing systems
US8741316 2014-06-03 Highly porous, recognitive polymer systems
US2012294869 2012-11-22 Methods for Treating Fatty Liver Disease
US2012078529 2012-03-29 DETERMINING THE SEVERITY OF 5-FLUOROURACIL OVERDOSE
US8067392 2011-11-29 Compositions and methods for treatment of mitochondrial diseases
US7915233 2011-03-29 Compositions and methods for treatment of mitochondrial diseases
US7884202 2011-02-08 Nucleobase Having Perfluoroalkyl Group and Process for Producing the Same

Uridine triacetate structure.svg

ChemSpider 2D Image | uridine triacetate | C15H18N2O9

Uridine triacetate.png

Uridine triacetate

  • Molecular FormulaC15H18N2O9
  • Average mass370.311 Da

ウリジントリアセタート

[(2R,3R,4R,5R)-3,4-diacetyloxy-5-(2,4-dioxopyrimidin-1-yl)oxolan-2-yl]methyl acetate

Uridine, 2′,3′,5′-triacetate
uridini triacetas
Vistogard [Trade name]
Xuriden [Trade name]
(2R,3R,4R,5R)-2-(acetoxymethyl)-5-(2,4-dioxo-3,4-dihydropyrimidin-1(2H)-yl)tetrahydrofuran-3,4-diyl diacetate
[(2R,3R,4R,5R)-3,4-bis(acetyloxy)-5-(2,4-dioxo-1,2,3,4-tetrahydropyrimidin-1-yl)oxolan-2-yl]methyl acetate
223-881-5 [EINECS]
2WP61F175M
4105-38-8 [RN]
9204
PN 401
RG 2133
TRIACETYL URIDINE

FDA APPROVED2015/9/4 . AS Xuriden

Uridine triacetate (INN),[1] formerly known as vistonuridine, is an orally active tri-acetylated prodrug of uridine[2] used:

  • in the treatment of hereditary orotic aciduria (brand name Xuriden /ˈzʊərədɛn/ ZOOR-ə-den);[3]
  • to treat patients following an overdose of chemotherapy drugs 5-fluorouracil (5-FU) or capecitabine regardless of the presence of symptoms, or who exhibit early-onset, severe or life-threatening toxicity affecting the cardiac or central nervous system, and/or early-onset, unusually severe adverse reactions (e.g., gastrointestinal toxicity and/or neutropenia) within 96 hours following the end of fluorouracil or capecitabine administration (brand name Vistogard).[4][5][6]

Uridine triacetate was developed, manufactured and distributed by Wellstat Therapeutics. Also, It was granted breakthrough therapy designation by FDA in 2015.

Uridine Triacetate is a synthetic uridine pro-drug that is converted to uridine in vivo. Uridine, a pyrimidine nucleotide, has been used in a variety of diseases including depressive disorders and inherited myopathies. (NCI04)

Uridine triacetate, formerly known as vistonuridine, is an orally active prodrug of the naturally occurring nucleoside uridine. It is used for the treatment of hereditary orotic aciduria (Xuriden), or for the emergency treatment of fluorouracil or capecitabine overdose or toxicity (Vistogard). It is provided in the prodrug form as uridine triacetate as this form delivers 4- to 6-fold more uridine into the systemic circulation compared to equimolar doses of uridine itself. When used for the treatment or prevention of toxicity associated with fluorouracil and other antimetabolites, uridine triacetate is utilized for its ability to compete with 5-fluorouracil (5-FU) metabolites for incorporation into the genetic material of non-cancerous cells. It reduces toxicity and cell-death associated with two cytotoxic intermediates: 5-fluoro-2′-deoxyuridine-5′-monophosphate (FdUMP) and 5-fluorouridine triphosphate (FUTP). Normally, FdUMP inhibits thymidylate synthase required for thymidine synthesis and DNA replication and repair while FUTPincorporates into RNA resulting in defective strands. As a result, these metabolites are associated with various unpleasant side effects such as neutropenia, mucositis, diarrhea, and hand–foot syndrome. Like many other neoplastic agents, these side effects limit the doses of 5-FU that can be administered, which also affects the efficacy for treatment. By pre-administering with uridine (as the prodrug uridine triacetate), higher doses of 5-FU can be given allowing for improved efficacy and a reduction in toxic side effects [A18578]. It can also be used as a rescue therapy if severe side effects present within 96 hours after initiation of therapy. Uridine triacetate is also used for the treatment of hereditary orotic aciduria, also known as uridine monophosphate synthase deficiency. This rare congenital autosomal recessive disorder of pyrimidinemetabolism is caused by a defect in uridine monophosphate synthase (UMPS), a bifunctional enzyme that catalyzes the final two steps of the de novo pyrimidine biosynthetic pathway. As a result of UMPS deficiency, patients experience a systemic deficiency of pyrimidine nucleotides, accounting for most symptoms of the disease. Additionally, orotic acid from the de novo pyrimidine pathway that cannot be converted to UMP is excreted in the urine, accounting for the common name of the disorder, orotic aciduria. Furthermore, orotic acid crystals in the urine can cause episodes of obstructive uropathy. When administered as the prodrug uridine triacetate, uridine can be used by essentially all cells to make uridine nucleotides, which compensates for the genetic deficiency in synthesis in patients with hereditary orotic aciduria. When intracellular uridine nucleotides are restored into the normal range, overproduction of orotic acid is reduced by feedback inhibition, so that urinary excretion of orotic acid is also reduced.

Uridine triacetate is an acetate ester that is uracil in which the three hydroxy hydrogens are replaced by acetate group. A prodrug for uridine, it is used for the treatment of hereditary orotic aciduria and for management of fluorouracil toxicity. It has a role as a prodrug, a neuroprotective agent and an orphan drug. It is a member of uridines and an acetate ester.

References

  1. ^ “International Nonproprietary Names for Pharmaceutical Substances (INN). Recommended International Nonproprietary Names: List 65” (PDF). World Health Organization. p. 92. Retrieved 12 March 2017.
  2. ^ “Uridine triacetate — DrugBank Page”. 12 March 2017.
  3. ^ “Xuriden (uridine triacetate) Oral Granules. Full Prescribing Information” (PDF). Wellstat Therapeutics Corporation. Gaithersburg, MD 20878. Retrieved 12 March 2017.
  4. ^ “Vistogard (uridine triacetate) Oral Granules. Full Prescribing Information” (PDF). Wellstat Therapeutics Corporation. Gaithersburg, MD 20878. Retrieved 12 March 2017.
  5. ^ “BTG Announces FDA Approval of Vistogard® (Uridine Triacetate) as Antidote to Overdose and Early Onset, Severe, or Life-Threatening Toxicities from Chemotherapy Drugs 5-Fluorouracil (5-FU) or Capecitabine”. BTG International Ltd. 11 December 2015. Retrieved 12 March 2017.
  6. ^ “Approved Drugs — Uridine Triacetate”. U.S. Food and Drug Administration. Retrieved 12 March 2017.

External links

Patents

  1. US7776838
  2. US5968914
  3. US6258795

FDA Orange Book Patents

FDA Orange Book Patents: 1 of 2 (FDA Orange Book Patent ID)
Patent 7776838
Expiration Aug 17, 2027
Applicant WELLSTAT THERAP
Drug Application N208159 (Prescription Drug: VISTOGARD. Ingredients: URIDINE TRIACETATE)
FDA Orange Book Patents: 2 of 2 (FDA Orange Book Patent ID)
Patent 6258795
Expiration Jul 10, 2019
Applicant WELLSTAT THERAP
Drug Application N208159 (Prescription Drug: VISTOGARD. Ingredients: URIDINE TRIACETATE)
Uridine triacetate
Uridine triacetate structure.svg
Clinical data
Trade names Vistogard, Xuriden
Routes of
administration
Oral granules
ATC code
Legal status
Legal status
Pharmacokinetic data
Metabolism Pyrimidine catabolic pathway
Onset of action Tmax = 2–3 hours
Elimination half-life 2–2.5 hours
Excretion Renal
Identifiers
CAS Number
PubChem CID
DrugBank
ChemSpider
UNII
KEGG
ECHA InfoCard 100.021.710 Edit this at Wikidata
Chemical and physical data
Formula C15H18N2O9
Molar mass 370.31 g·mol−1
3D model (JSmol)

////////////Uridine triacetate, ウリジントリアセタート  , FDA 2015, breakthrough therapy designation ,

//////////174105-38-8Priority review drug , Orphan drug, FDA 2015,  Vistogard, uridine triacetate, fast track designations, PN-401, RG-2133,  TAU, XURIDEN
CC(=O)OC[C@H]1O[C@H]([C@H](OC(C)=O)[C@@H]1OC(C)=O)N1C=CC(=O)NC1=O

FDA approves new oral therapy to treat ALK-positive lung cancer


12/11/2015 01:03 PM EST
The U.S. Food and Drug Administration today approved Alecensa (alectinib) to treat people with advanced (metastatic) ALK-positive non-small cell lung cancer (NSCLC) whose disease has worsened after, or who could not tolerate treatment with, another therapy called Xalkori (crizotinib).


December 11, 2015

Release

The U.S. Food and Drug Administration today approved Alecensa (alectinib) to treat people with advanced (metastatic) ALK-positive non-small cell lung cancer (NSCLC) whose disease has worsened after, or who could not tolerate treatment with, another therapy called Xalkori (crizotinib).

Lung cancer is the leading cause of cancer death in the United States, with an estimated 221,200 new diagnoses and 158,040 deaths in 2015, according to the National Cancer Institute. An ALK (anaplastic lymphoma kinase) gene mutation can occur in several different types of cancer cells, including lung cancer cells. ALK gene mutations are present in about 5 percent of patients with NSCLC. In metastatic cancer, the disease spreads to new parts of the body. In ALK-positive NSCLC metastatic patients, the brain is a common place for the disease to spread.

“Today’s approval provides a new therapy for a group of patients who would have few treatment options once their disease no longer responds to treatment with Xalkori,” said Richard Pazdur, M.D., director of the Office of Hematology and Oncology Products in the FDA’s Center for Drug Evaluation and Research. “In addition to the primary effect on tumors in the lung, Alecensa clinical trials provide evidence of an effect on tumors that had spread to the brain, which is an important effect for clinicians to understand.”

Alecensa is an oral medication that blocks the activity of the ALK protein, which may prevent NSCLC cells from growing and spreading.

The safety and efficacy of Alecensa were studied in two single-arm clinical trials of patients with metastatic ALK-positive NSCLC whose disease was no longer controlled by treatment with Xalkori. Study participants received Alecensa twice daily to measure the drug’s effect on their lung cancer tumors. In the first study, 38 percent of participants experienced a partial shrinkage of their NSCLC tumors, an effect that lasted for an average of 7.5 months. In the second study, 44 percent of participants experienced a partial shrinkage of their NSCLC tumors, lasting for an average of 11.2 months. The trials also examined Alecensa’s effect on individuals’ brain metastases, a common occurrence in this population. Sixty-one percent of participants in the two trials who had measurable brain metastases experienced a complete or partial reduction in their brain tumors, lasting an average of 9.1 months.

The most common side effects of Alecensa are fatigue, constipation, swelling (edema) and muscle pain (myalgia). Alecensa may cause serious side effects, including liver problems, severe or life-threatening inflammation of the lungs, very slow heartbeats and severe muscle problems. Treatment with Alecensa may cause sunburn when patients are exposed to sunlight.

Alecensa was approved using the accelerated approval regulatory pathway, which allows the FDA to approve products for serious or life-threatening diseases based on evidence that the product has an effect on an outcome that is reasonably likely to predict clinical benefit. In the case of Alecensa, the tumor response to treatment, along with the duration of response, provided this evidence. Under the accelerated approval requirements, a confirmatory study is required to verify and describe the clinical benefit of Alecensa.

The FDA granted the Alecensa application breakthrough therapy designation and priority review status. These are distinct programs intended to facilitate and expedite the development and review of certain new drugs in light of their potential to benefit patients with serious or life-threatening conditions. Alecensa also received orphan drug designation, which provides incentives such as tax credits, user fee waivers and eligibility for exclusivity to assist and encourage the development of drugs for rare diseases.

Alecensa is marketed by Genentech, based in San Francisco, California. Xalkori is marketed by Pfizer, based in New York, New York.


Synthesis


Read also

https://newdrugapprovals.org/2014/07/08/japan-first-to-approve-alectinib-%E3%82%A2%E3%83%AC%E3%82%AF%E3%83%81%E3%83%8B%E3%83%96-%E5%A1%A9%E9%85%B8%E5%A1%A9-af-802-for-alk-nsclc/


/////////////////

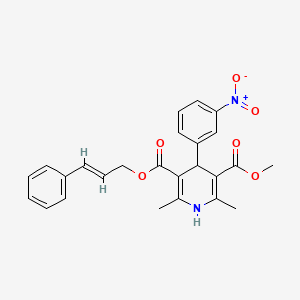
Pranidipine


str1

File:Pranidipine structure.svg

Pranidipine , OPC-13340, FRC 8411

Acalas®

NDA Filing in Japan

A calcium channel blocker potentially for the treatment of angina pectoris and hypertension.

CAS No. 99522-79-9

  • Molecular FormulaC25H24N2O6
  • Average mass448.468
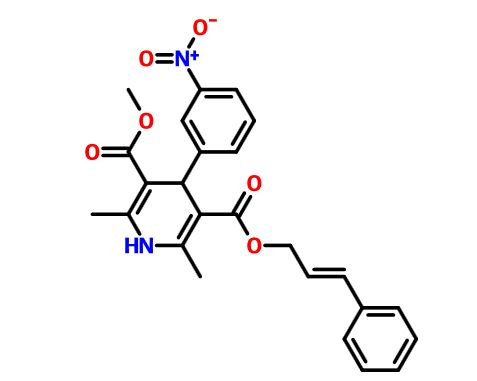
methyl (2E)-3-phenylprop-2-en-1-yl 2,6-dimethyl-4-(3-nitrophenyl)-1,4-dihydropyridine-3,5-dicarboxylate
Methyl-(2E)-3-phenyl-2-propen-1-yl-2,6-dimethyl-4-(3-nitrophenyl)-1,4-dihydro-3,5-pyridindicarboxylat  (E)-Cinnamyl methyl (±)-1,4-dihydro-2,6-dimethyl-4-(m-nitrophenyl)-3,5-pyridinedicarboxylate
Methyl cinnamyl 2,6-dimethyl-4-(3-nitrophenyl)-1,4-dihydro-3,5-pyridinedicarboxylate
trans-Cinnamyl methyl 4-(3-nitrophenyl)-2,6-dimethyl-1,4-dihydropyridine-3,5-dicarboxylate
3,5-Pyridinedicarboxylic acid, 1,4-dihydro-2,6-dimethyl-4-(3-nitrophenyl)-, methyl (2E)-3-phenyl-2-propen-1-yl ester

Pranidipine is a calcium channel blocker. It is a long acting calcium channel antagonist of the dihydropyridine group.[1]


PATENT

EP 0173126

http://www.google.com/patents/EP0173126A1?cl=en

PAPER

Der Pharmacia Sinica, 2014, 5(1):11-17

pelagiaresearchlibrary.com/der-pharmacia-sinica/vol5-iss1/DPS-2014-5-1-11-17.pdf


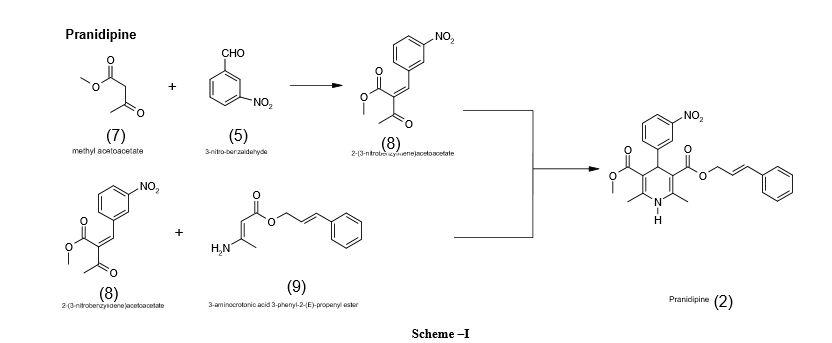
str1

 CLICK ON IMAGE FOR CLEAR VIEW

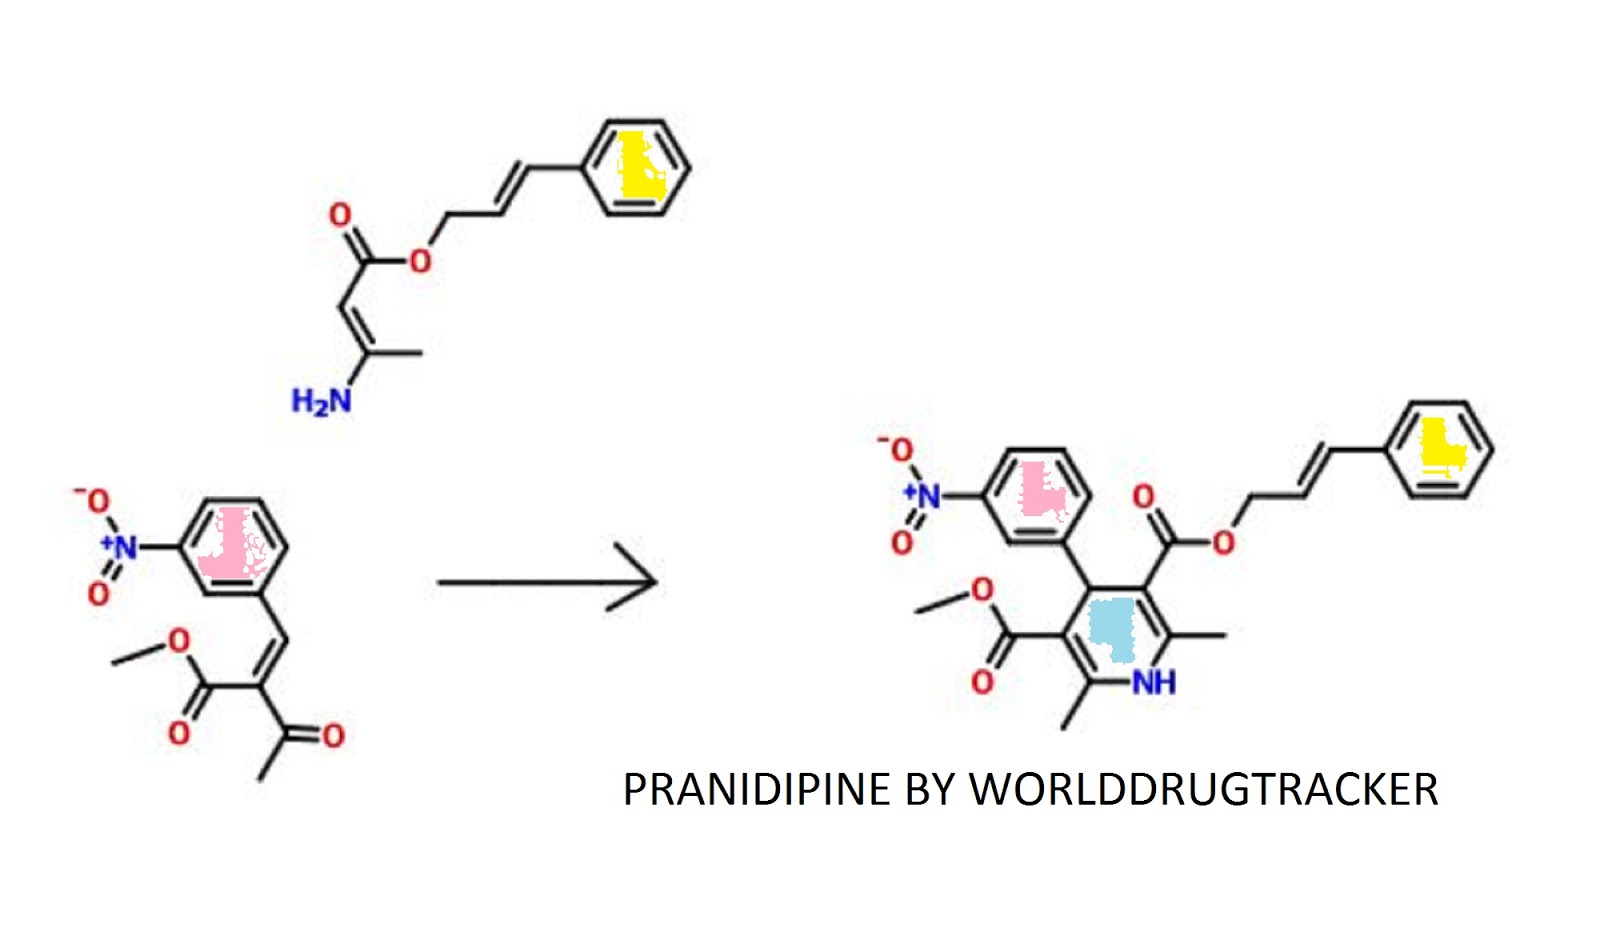
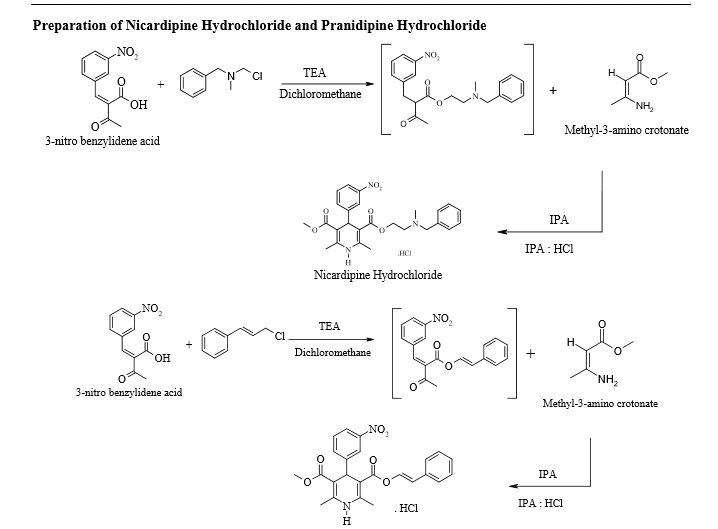
Preparation of Pranidipine Hydrochloride(2):
To a suspension of (Z)-2-(3-nitrobenzylidene)-3-oxobutanoic acid(3) (1.2 kg, 5.10 mol) in dichloromethane (6 L)
was added triethylamine(0.77 kg, 7.65 mol) and cinnamyl chloride (0.85 kg, 5.61 mol). The reaction mixture was
heated to 45°C and maintained for 2 hrs. The suspension was cooled to 25 to 30°C and washed with 2.4 Lof DM
water. DCM layer was separated and concentrated under vacuum below 40°C. The concentrated mass was dissolved
in 7.2 L isopropyl alcohol and methyl-3-amino crotonate (0.52 kg, 4.5mol) was added to it. Temperatureof reaction
mixture was slowly raised to 70°C and maintained for 8 hours. Reaction mass was concentrated under vacuum
below 40°C.To the crude residue, ethyl acetate-HCl(0.28 kg, 7.6 mol) was added and the reaction mixture was
stirred for 24 hours at 25°C-30°C. Reaction mixturewas filtered and the solid residue was dried under
vacuum toafford 1.6 kg of Pranidipine hydrochloride (2)in 85% yield with 98 % purity.
1H-NMR(DMSO):
δ2.29 (s, 3H),2.32 (s, 3H), 3.55 (s, 3H), 4.60-4.74 (m, 2H), 5.04(s, 1H), 6.26-6.33 (m, 1H), 6.50 (d, 1H), 7.24-7.3
8 (m, 5H), 7.53(t, 1H), 7.63 (d,1H), 7.98-8.01 (m, 1H), 9.08 (brs, 1H)
 
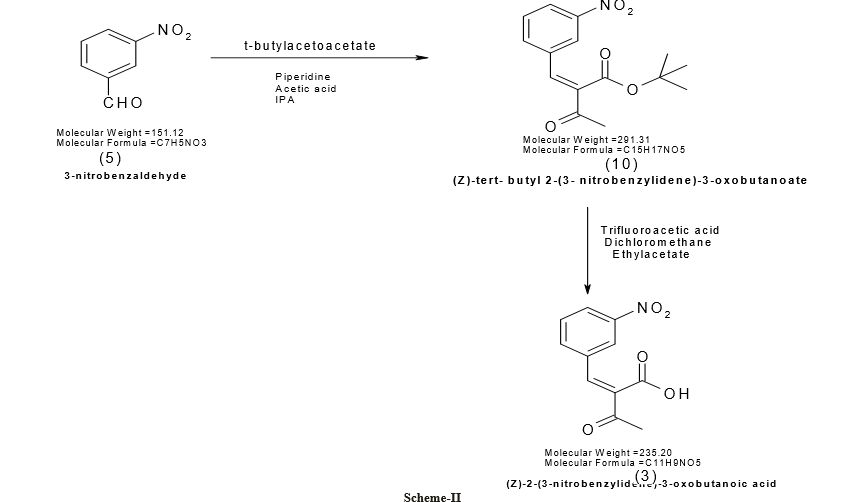
Preparation of (Z)-2-(3-nitrobenzylidene)-3-oxobutanoic acid(3):
To a suspension of (Z)-t-butyl 2-(3-nitrobenzylidene)-3-oxobutanoate(10) (1.5 kg, 5.14 mol) in dichloromethane
(7.5 L) was added trifluoroacetic acid (1.76 kg, 15.44 mol) and reaction mass was stirred at 25°C to 30°C for 24 hrs.
The reaction mass was concentrated under vacuum below 40°C and stripped with toluene. The concentratedmass
was dissolved in 4.5 L toluene and the solution wasstirred for 8 hours at 25°C to 30°C. Reaction mixture was
filtered and solid washed with toluene and dried at35°C to 40°C to give 1.152 kg of (Z)-2-(3-nitrobenzylidene)-3-
oxobutanoic acid(3) in 96 % yield. M.P: 120°C; Mol.Wt: 235.20; Mol.Formula: C11H9NO5;1H-NMR(DMSO):
δ2.46 (s, 3H), 7.76-7.83 (m, 2H), 8.02 (d, 1H), 8.28-8.31 (dd, 1H), 8.51 (s, 1H), 13.63 (brs, 1H).Anal.Calcd for
C11H10NO5 : C, 55.93; H, 4.27; N, 5.93. Found: C, 56.19;H, 4.09; N, 6.27
Preparation of (Z)-tertiary- butyl 2-(3-nitrobenzylidene)-3-oxobutanoate(10):
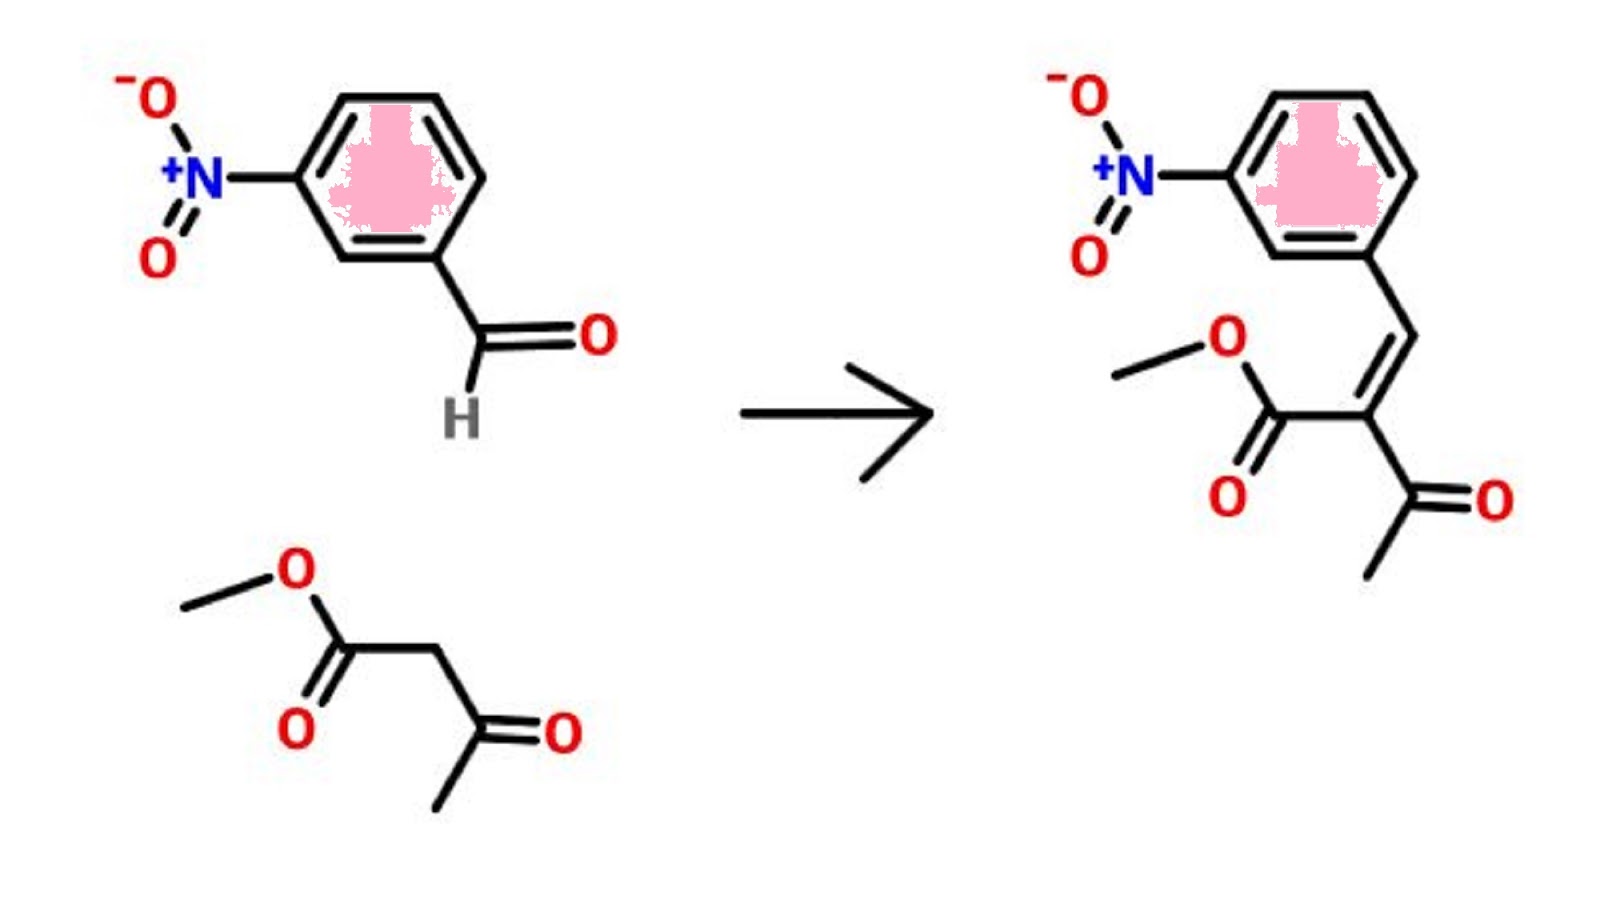
To a suspension of 3-nitrobenzaldehyde(5) (1 kg, 6.61 mol) in isopropyl alcohol (6 L) was addedt-butylacetoacetate (1.14 kg, 7.27 mol),piperidine (0.12 kg, 1.32 mol) and acetic acid (0.79 kg, 1.32 mol). The reactionmass was stirred at 25°C to 30°C for 6 hrs. The suspension was cooled to -5 to 0°C, filtered, residuewashed withisopropyl alcohol and dried at 35°C to 40°C to give
1.750 kg of (Z)-t-butyl 2-(3-nitrobenzylidene)-3-oxobutanoate(10)in 91% yield; M.P: 80°C; Mol. Wt: 291.31; Mol.Formula: C15H17NO5
1H-NMR(CDCl3):
δ1.55(s, 9H), 2.44 (s, 3H), 7.50 (s, 1H),7.59 (t, 1H),7.80 (d, 1H), 8.24- 8.27 (dd,J=1H),δ8.41 (t, 1H).
str1
str1
  CLICK ON IMAGE FOR CLEAR VIEW
Patent Submitted Granted
Process for the preparation of 1,4 – dihydropyridines and novel 1,4-dihydropyridines useful as therapeutic agents [US2003230478] 2003-12-18
Advanced Formulations and Therapies for Treating Hard-to-Heal Wounds [US2014357645] 2014-08-19 2014-12-04
METHODS OF TREATING CARDIOVASCULAR AND METABOLIC DISEASES [US2014322199] 2012-08-06 2014-10-30
Protein Carrier-Linked Prodrugs [US2014323402] 2012-08-10 2014-10-30
sGC STIMULATORS [US2014323448] 2014-04-29 2014-10-30
TREATMENT OF ARTERIAL WALL BY COMBINATION OF RAAS INHIBITOR AND HMG-CoA REDUCTASE INHIBITOR [US2014323536] 2012-12-07 2014-10-30
Agonists of Guanylate Cyclase Useful For the Treatment of Gastrointestinal Disorders, Inflammation, Cancer and Other Disorders [US2014329738] 2014-03-28 2014-11-06
METHODS, COMPOSITIONS, AND KITS FOR THE TREATMENT OF CANCER [US2014335050] 2012-05-25 2014-11-13
ROR GAMMA MODULATORS [US2014343023] 2012-09-18 2014-11-20
High-Loading Water-Soluable Carrier-Linked Prodrugs [US2014296257] 2012-08-10 2014-10-02


Pranidipine.png

Publication Number Publication Date IPCR Assignee/Applicant Structure hits Tools
1.

US-20150342954-A1

2015-12-03
2-BENZYL, 3-(PYRIMIDIN-2-YL) SUBSTITUTED PYRAZOLES USEFUL AS SGC STIMULATORS
COC(=O)C1=C(C)NC(C)=C(C1C1=CC(=CC=C1)[N+]([O-])=O)C(=O)OC\C=C\C1=CC=CC=C1
2.

EP-2558474-B1

2015-11-25
2, 4-PYRIMIDINEDIAMINE COMPOUNDS AND PRODRUGS THEREOF AND THEIR USES
EN
COC(=O)C1=C(C)NC(C)=C(C1C1=CC(=CC=C1)[N+]([O-])=O)C(=O)OC\C=C\C1=CC=CC=C1
3.

US-20150307580-A1

2015-10-29
OXYNTOMODULIN ANALOGS
COC(=O)C1=C(C)NC(C)=C(C1C1=CC(=CC=C1)[N+]([O-])=O)C(=O)OC\C=C\C1=CC=CC=C1
4.

US-20150305974-A1

2015-10-29
METHODS AND DEVICES FOR TREATING HYPERTENSION
COC(=O)C1=C(C)NC(C)=C(C1C1=CC(=CC=C1)[N+]([O-])=O)C(=O)OC\C=C\C1=CC=CC=C1
5.

WO-2015164658-A1

2015-10-29
METHODS AND DEVICES FOR TREATING HYPERTENSION
EN
COC(=O)C1=C(C)NC(C)=C(C1C1=CC(=CC=C1)[N+]([O-])=O)C(=O)OC\C=C\C1=CC=CC=C1
6.

EP-2527360-B1

2015-10-28
Agonists of guanylate cyclase useful for the treatment of gastrointestinal disorders, inflammation, cancer and other disorders
EN
COC(=O)C1=C(C)NC(C)=C(C1C1=CC(=CC=C1)[N+]([O-])=O)C(=O)OC\C=C\C1=CC=CC=C1
7.

WO-2015157471-A1

2015-10-15
INOS-INHIBITORY COMPOSITIONS AND THEIR USE AS BREAST CANCER THERAPEUTICS
EN
COC(=O)C1=C(C)NC(C)=C(C1C1=CC(=CC=C1)[N+]([O-])=O)C(=O)OC\C=C\C1=CC=CC=C1
8.

US-20150284411-A1

2015-10-08
NOVEL AZABENZIMIDAZOLE HEXAHYDROFURO[E,2-B]FURAN DERIVATIVES
COC(=O)C1=C(C)NC(C)=C(C1C1=CC(=CC=C1)[N+]([O-])=O)C(=O)OC\C=C\C1=CC=CC=C1
9.

US-20150283202-A1

2015-10-08
AGONISTS OF GUANYLATE CYCLASE USEFUL FOR THE TREATMENT OF HYPERCHOLESTEROLEMIA, ATHEROSCLEROSIS, CORONARY HEART DISEASE, GALLSTONE, OBESITY AND OTHER CARDIOVASCULAR DISEASES
COC(=O)C1=C(C)NC(C)=C(C1C1=CC(=CC=C1)[N+]([O-])=O)C(=O)OC\C=C\C1=CC=CC=C1
10.

US-9150512-B2

2015-10-06
Tricyclic lactam derivatives as 11-beta hydroxysteroid dehydrogenase inhibitors
COC(=O)C1=C(C)NC(C)=C(C1C1=CC(=CC=C1)[N+]([O-])=O)C(=O)OC\C=C\C1=CC=CC=C1

References

Jin Yang, Keisuke Fukuo, Shigeto Morimoto, Tadaaki Niinobu, Toshimitsu Suhara, Toshio Ogihara (2000). “Pranidipine Enhances the Action of Nitric Oxide Released From Endothelial Cells”. Hypertension 35: 82–85. doi:10.1161/01.hyp.35.1.82.


http://pelagiaresearchlibrary.com/der-pharmacia-sinica/vol5-iss1/DPS-2014-5-1-11-17.pdf………NICARDIPINE

Pranidipine
Pranidipine structure.svg
Names
IUPAC name

methyl (2E)-phenylprop-2-en-1-yl 2,6-dimethyl-4-(3-nitrophenyl)-1,4-dihydropyridine-3,5-dicarboxylate
Other names

2,6-dimethyl-4-(3-nitrophenyl)-1,4-dihydropyridine-3,5-dicarboxylic acid O5-methyl O3-[(E)-3-phenylprop-2-enyl] ester
Identifiers
99522-79-9 Yes
ChEMBL ChEMBL1096842 
ChemSpider 4940726 
Jmol interactive 3D Image
MeSH C048161
PubChem 6436048
UNII 9DES9QVH58 Yes
Properties
C25H24N2O6
Molar mass 448.46786


//////////

CC1=C(C(C(=C(N1)C)C(=O)OCC=CC2=CC=CC=C2)C3=CC(=CC=C3)[N+](=O)[O-])C(=O)OC

see dipine series………..http://organicsynthesisinternational.blogspot.in/p/dipine-series.html

Nilvadipine – Wikipedia, the free encyclopedia

manidipine

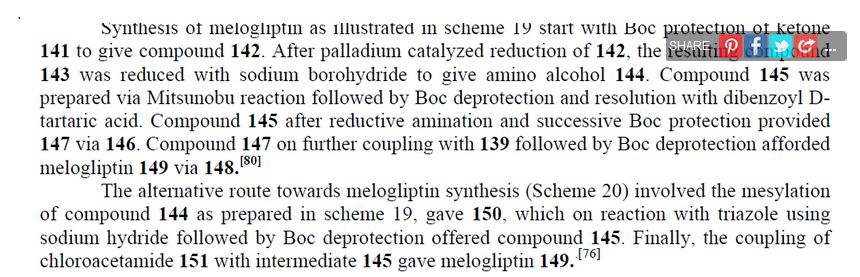
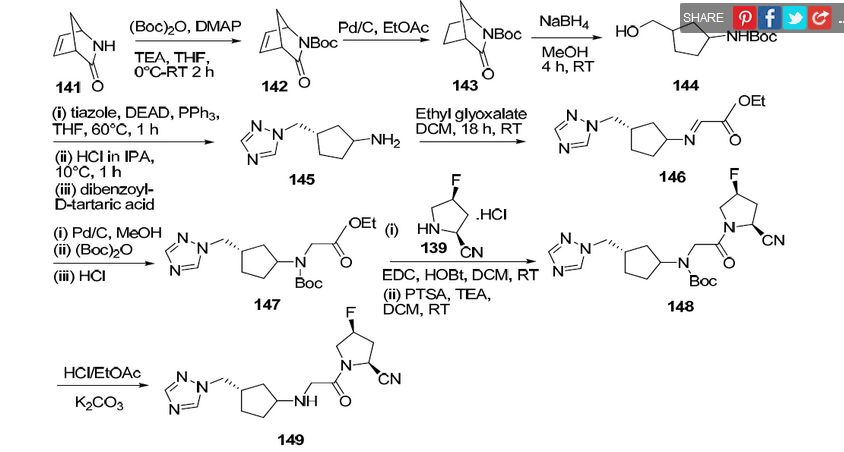
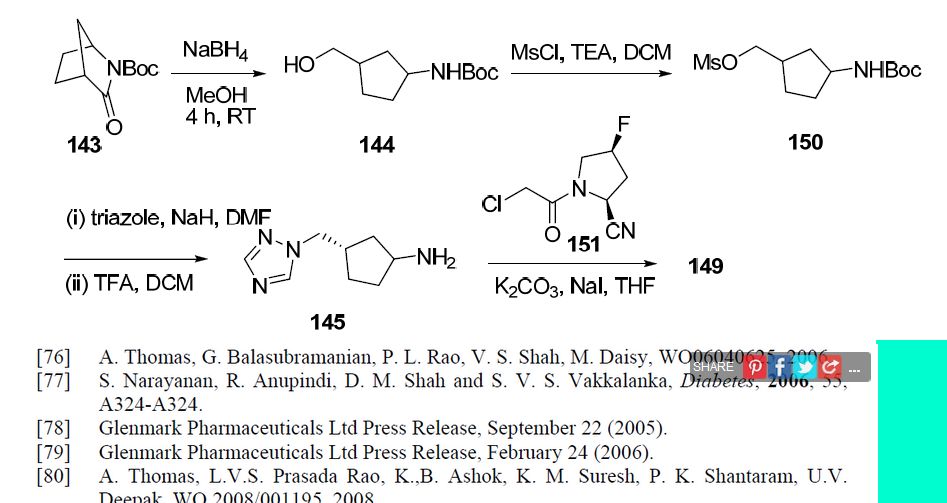
MELOGLIPTIN


Melogliptin

Phase III

A DP-IV inhibitor potentially for treatment of type II diabetes.

EMD-675992; GRC-8200

CAS No. 868771-57-7

4-fluoro-1-[2-[[(1R,3S)-3-(1,2,4-triazol-1-ylmethyl)cyclopentyl]amino]acetyl]pyrrolidine-2-carbonitrile
4(S)-Fluoro-1-[2-[(1R,3S)-3-(1H-1,2,4-triazol-1-ylmethyl)cyclopentylamino]acetyl]pyrrolidine-2(S)-carbonitrile
Note………The views expressed are my personal and in no-way suggest the views of the professional body or the company that I represent
MELOGLIPTIN

GRC-8200, a dipeptidyl peptidase IV inhibitor (DPP-IV), is currently undergoing phase II clinical trials at Glenmark Pharmaceuticals and Merck KGaA for the treatment of type 2 diabetes. In 2006, the compound was licensed by Glenmark Pharmaceuticals to Merck KGaA in Europe, Japan and N. America for the treatment of type 2 diabetes, however, these rights were reaquired by Glenmark in 2008.
str1
str1


DISCLAIMER…….The views expressed are my personal and in no-way suggest the views of the professional body or the company that I represent

/////////

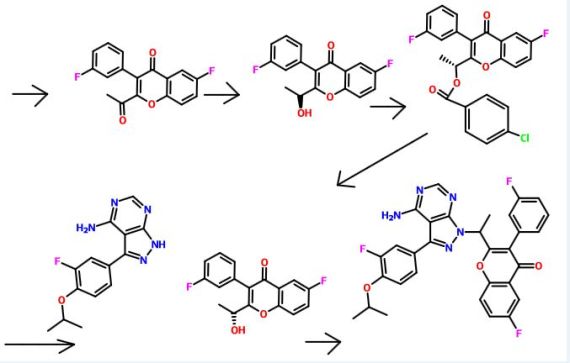
Umbralisib, TGR-1202, a Phosphoinositide-3 kinase delta inhibitor, Rhizen Pharmaceuticals S.A./TG Therapeutics


rp-5264.png
TGR 1202, TGR-1202-101, RP 5264, Umbralisib
AK173784;
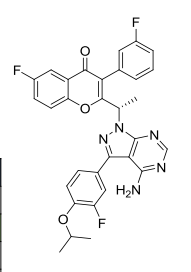
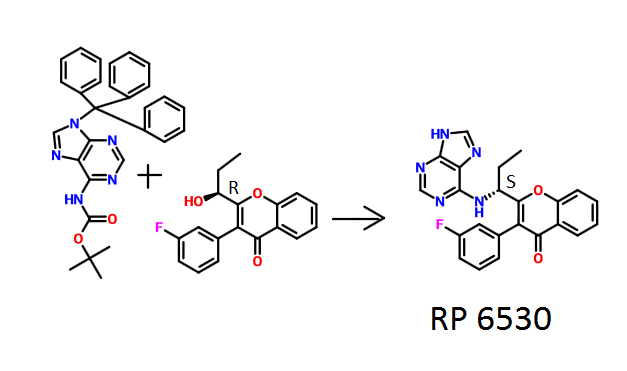
(S)-2-(1-(4-amino-3-(3-fluoro-4-isopropoxyphenyl)-1H-pyrazolo[3,4-d]pyrimidin-1-yl)ethyl)-6-fluoro-3-(3-fluorophenyl)-4H-chromen-4-one
(S)-2-(l-(4-amino-3-(3-fluoro-4-isopropoxyphenyl)-lH-pyrazolo[3,4-d]pyrimidin-l-yl)ethyl)-3-(3-fluorophenyl)-4H-chromen-4-one,
2-[(1S)-1-[4-amino-3-(3-fluoro-4-propan-2-yloxyphenyl)pyrazolo[3,4-d]pyrimidin-1-yl]ethyl]-6-fluoro-3-(3-fluorophenyl)chromen-4-one
 CAS TOSYLATE 1532533-72-4 Umbralisib tosylate
CAS 1532533-67-7, 1514919-95-9
Molecular Formula: C31H24F3N5O3
Molecular Weight: 571.54917 g/mol

RP-5307
TGR-1202
TGR-1202 PTSA
FU8XW5V3FS (UNII code)
RP-5264 (free base)

TGR‐1202, a next generation PI3K-δ delta inhibitor. TGR-1202 (RP-5264) is a highly specific, orally available, PI3K delta inhibitor, targeting the delta isoform with nanomolar potency and several fold selectivity over the alpha, beta, and gamma isoforms of PI3K.

TG Therapeutics, under license from Rhizen Pharmaceuticals, is developing TGR-1202 (structure shown; formerly RP-5264), a lead from a program of PI3K delta inhibitors, for the potential oral treatment of hematological cancers including Hodgkin lymphoma, non-Hodgkin lymphoma (NHL), chronic lymphocytic leukemia (CLL), B-cell lymphoma and mantle cell lymphoma (MCL)

Incozen Therapeutics Pvt Ltd

TG Therapeutics
TGR-1202 potential to perform as the best PI3K inhibitor in its class and the possible superiority of TG-1101 over Rituxan®.

Rhizen Pharmaceuticals S.A.
Description Phosphoinositide 3-kinase (PI3K) delta inhibitor

Leukemia, chronic lymphocytic  PHASE 3, TG Therapeutics

Orphan Drug

Umbralisib is a novel phosphatidylinositol 3-kinase delta (PI3Kdelta) inhibitor under development at TG Therapeutics in phase III clinical trials, in combination with ublituximab, for the treatment of chronic lymphocytic leukemia (CLL) and for the treatment of diffuse large B-cell lymphoma (DLBCL). The company refers to the combination regimen of ublituximab and TGR-1202 as TG-1303. The drug is also in phase II clinical development for the oral treatment of hematologic malignancies, as a single agent or in combination therapy. Phase I clinical trials are ongoing in patients with select relapsed or refractory solid tumors, such as adenocarcinoma of the pancreas, adenocarcinoma of the colon, rectum, gastric and GE junction cancer, and GI Stromal Tumor (GIST).

In 2016, orphan drug designation was assigned to the compound in the U.S. for the treatment of CLL. In 2017, additional orphan drug designation was granted in the U.S. for the treatment of CLL and DLBCL, in combination with ublituximab.

Originated by Rhizen Pharmaceuticals, the product was jointly developed by Rhizen Pharmaceuticals and TG Therapeutics since 2012. In 2014, exclusive global development and commercialization rights (excluding India) were licensed to TG Therapeutics.

CLINICAL TRIALS……….https://clinicaltrials.gov/search/intervention=TGR-1202

B-cell lymphoma; Chronic lymphocytic leukemia; Hematological neoplasm; Hodgkins disease; Mantle cell lymphoma; Non-Hodgkin lymphoma

Phosphoinositide-3 kinase delta inhibitor
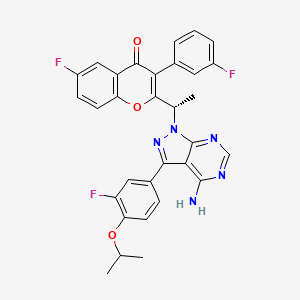
rp-5264.png

SYNTHESIS

str1

str1

Rhizen Pharmaceuticals Announces Out-licensing Agreement for TGR-1202, a Novel Next Generation PI3K-delta Inhibitor

Rhizen to receive upfront payment of $8.0 million — Rhizen to retain global manufacturing and supply rights — Rhizen to retain development and commercialization for India

Rhizen to retain development and commercialization for India

| Source: Rhizen Pharmaceuticals SA

La Chaux-de-Fonds, Switzerland, Sept. 23, 2014 (GLOBE NEWSWIRE) — Rhizen Pharmaceuticals S.A. today announced an out-licensing agreement for TGR-1202, a novel next generation PI3K-delta inhibitor. TG Therapeutics exercised its option for early conversion to a licensing agreement from a 50:50 joint venture partnership.

In exchange for this licensing agreement, TG Therapeutics will pay Rhizen an upfront payment of $8.0 million ($4.0 million in cash and $4.0 million in TG Therapeutics common stock).  In addition to the upfront payment, Rhizen will be eligible to receive regulatory filing, approval and sales based milestones in the aggregate of approximately $240 million, and tiered royalties based on net sales.

Swaroop Vakkalanka, Ph.D. and President of Rhizen stated, “We are extremely happy and take pride in discovering a novel, next generation, once-daily PI3K-delta inhibitor under active development led by TG Therapeutics.  We are encouraged by the progress of TRG-1202 to date, and the speed at which TG Therapeutics is developing the asset in various hematological malignancies.  We look forward to the day this novel drug reaches cancer patients in need of new and safe therapies.”

About Rhizen Pharmaceuticals S.A.:

Rhizen Pharmaceuticals is an innovative, clinical-stage biopharmaceutical company focused on the discovery and development of novel therapeutics for the treatment of cancer, immune and metabolic disorders.  Since its establishment in 2008, Rhizen has created a diverse pipeline of proprietary drug candidates targeting several cancers and immune associated cellular pathways.  Rhizen is headquartered in La-Chaux-de-Fonds, Switzerland.  For additional information, please visit Rhizen’s website, www.rhizen.com.


TGR-1202.with Idelalisib and IPI-145 (left to right) for comparison.
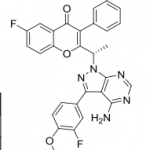
TGTX structureIdelalisib StrutureIPI-145 Structure

IPI 145

PATENTS

WO 2011055215

http://www.google.com/patents/WO2011055215A2?cl=en

PATENT

WO 2015181728

https://patentscope.wipo.int/search/en/detail.jsf?docId=WO2015181728

TGR-1202, chemically known as (S)-2-(l-(4-amino-3-(3-fluoro-4-isopropoxyphenyl)-lH-pyrazolo[3,4-d]pyrimidin-l-yl)ethyl)-3-(3-fluorophenyl)-4H-chromen-4-one, has the following chemical structure:

Example 1: Preparation of the PTSA Salt of TGR-1202 (Form A)

7100 g of TGR-1202 was charged in a reactor containing 56.8 litres of acetone and stirred at ambient temperature. 4680 g of p-toluene sulphonic acid was added and the reaction mixture was heated at a temperature of 60-65° C for about 6 hours. The solvent was removed by distillation under reduced pressure to obtain a wet residue. The wet residue was degassed and allowed to cool to < 20° C. Approximately 142 litres of diethyl ether was then added and the resulting mixture was stirred overnight, then filtered to obtain a solid mass which was washed with diethyl ether and dried in vacuo to yield a solid mass. The solid mass was re-suspended in diethyl ether, stirred for 6 hours, and then filtered to yield a solid mass which was subsequently dissolved in 56.8 litres of acetone, filtered through a HiFlow bed, and concentrated under reduced pressure. The resulting residue mass was stirred with water overnight, then filtered and vacuum dried to yield 6600 g of the PTSA salt of TGR-1202. HPLC: 99.21% and chiral purity of 99.64:0.36 (S:R).

Example 2: Preparation of the PTSA Salt of TGR-1202 (Form B)

1000 g of TGR-1202 was charged in a reactor containing 8 litres of acetone and stirred at ambient temperature. 666 g of p-toluene sulphonic acid was then added and the reaction mixture was heated at a temperature of 60-65 °C for about 6 hours. The solvent was removed by distillation under reduced pressure to obtain a wet residue. The wet residue was degassed and allowed to cool to < 20° C. Approximately 20 litres of diethyl ether was added and the resulting mixture was stirred overnight, then filtered to obtain a solid mass which was washed with diethyl ether and dried in vacuo to yield a solid mass which was then vacuum dried to yield 1150 g of the PTSA salt of TGR-1202. HPLC: 99.33% and chiral purity: 99.61:0.39 (S:R).

Table 1 lists the XRPD pattern peaks and relative peak intensities for the products of Examples 1 and 2.

TABLE 1

The tablet composition comprising a PTSA salt of TGR-1202 prepared according to Example 2 exhibited a Cmax about 2.5 fold and an area under the curve (AUC) about 1.9 fold greater than that of the tablet composition comprising a PTSA salt of TGR-1202 prepared according to Example 1. The results are provided in Table 8 below.

TABLE 8

PATENT

WO 2014071125

http://www.google.com/patents/WO2014071125A1?cl=en

formula (A) that is a ΡΒΚδ selective inhibitor,

(A)

Synthesis of Compound of Formula A

Unless otherwise stated, purification implies column chromatography using silica gel as the stationary phase and a mixture of petroleum ether (boiling at 60-80°C) and ethyl acetate or dichloromethane and methanol of suitable polarity as the mobile phases. The term “RT” refers to ambient temperature (25-28°C).

Intermediate 1 : 2-( l-bromoethyl)-6-fluoro-3-f3-fluorophenyl)-4H-chromen-4-one

Step-1 [l-(5-Fluoro-2-hydroxyphenyl)-2-(3-fluorophenyl)ethanone]: 3- Fluorophenylacetic acid (7.33 g, 47.56 mmoles) was dissolved in 25 ml dichloromethane. To this mixture, oxalylchloride (7.54 g, 59.46 mmoles) and DMF (3 drops) were added at 0°C and stirred for 30 min. The solvent was evaporated and dissolved in 25 ml dichloromethane. To this mixture, 4-fluoroanisole (5.00 g, 39.64 mmoles) was added and cooled to 0°C. At 0°C A1C13 (7.95 g, 59.46 mmoles) was added and the reaction mixture was warmed to RT and stirred for 12 hours. The reaction mixture was quenched by the addition of 2N HC1, extracted with ethyl acetate, dried over sodium sulphate and concentrated. The crude product was purified by column chromatography with ethyl acetate :petroleum ether to afford the title compound as colorless solid (4.5 g, 45% yield). 1H-NMR (δ ppm, DMSO-D6, 400 MHz): δ 11.34 (s, 1H), 7.75 (dd, J=9.4, 3.1 Hz, 1H), 7.42 (m, 2H), 7.12 (m, 3H), 7.05 (dd, J=9.0, 4.5 Hz, 1H), 4.47 (s, 2H).

Step-2 [2-Ethyl-6-fiuoro-3-(3-fluorophenyl)-4H-chromen-4-one]: l-(5-Fluoro-2- hydroxyphenyl)-2-(3-fluorophenyl)ethanone obtained from Step-1 (3.00 g, 12.08 mmoles) was placed in a round bottom flask and to this triethylamine (25 ml) and propionic anhydride (4.92 g, 37.82 mmoles) were added, and the mixture was refluxed for 24 hours. After cooling to RT, the reaction mixture was acidified by the addition of IN HC1 solution, extracted with ethyl acetate, washed with sodium bicarbonate solution, dried with sodium sulphate and concentrated. The crude product was purified by column chromatography with ethyl acetate :petroleum ether to afford the title compound as off-yellow solid (1.80 g, 52% yield). 1H-NMR (δ ppm, DMSO-D6, 400 MHz): δ 7.80 (m, 1H), 7.76 (m, 2H), 7.51 (dd, J=8.0, 6.4 Hz), 7.22 (m, 1H), 7.18 (m, 2H), 2.56 (q, J=7.6 Hz, 2H), 1.20 (t, J=7.6 Hz, 3H).

Step-3: To a solution of 2-Ethyl-6-fluoro-3-(3-fluorophenyl)-4H-chromen-4-one obtained from Step-2 (1.80 g, 6.28 mmoles) in carbon tetrachloride (20 ml), N- bromosuccinimide (1.11 g, 6.28 mmoles) was added and heated to 80°C. Azobisisobutyronitrile (10 mg) was added to the reaction mixture at 80°C. After 12 hours, the reaction mixture was cooled to RT, diluted with dichloromethane and washed with water. The organic layer was dried over sodium sulphate and concentrated under reduced pressure to afford the crude title compound as yellow solid (1.25 g, 55% yield). 1H-NMR (δ ppm, DMSO-D6, 400 MHz): δ 7.91 (dd, J=9.2, 4.3 Hz, 1H), 7.81 (dt, j=8.2, 2.8 Hz, 1H), 7.74 (dd, J=8.3, 3.1 Hz, 1H), 7.57 (m, 1H), 7.32 (dt, J=8.5, 2.4 Hz, 1H), 7.19 (m, 2H), 5.00 (q, J=6.8 Hz, 1H), 1.97 (d, J=6.8 Hz, 3H).

Intermediate 2: 6-fluoro-3-f3-fluorophenyl)-2-fl-hvdroxyethyl)-4H-chromen-4-one

To a solution of Intermediate 1 (15.0 g, 40.84 mmol) in DMSO (150 ml), n-butanol (7.5 ml) was added and heated to 120°C for 3 hours. The reaction mixture was cooled to RT, quenched with water and extracted with ethyl acetate. The organic layer was dried over sodium sulphate and concentrated under reduced pressure. The crude product was purified by column chromatography with ethyl acetate: petroleum ether to afford the title compound as an off-white solid (7.90 g, 64%). 1H-NMR (δ ppm, CDC13, 400 MHz): 7.85 (dd, J = 8.1, 3 Hz, 1H), 7.54 (dd, J = 9.2, 4.2 Hz, 1H), 7.47-7.37 (m, 2H), 7.15-6.98 (m, 3H), 4.74 (quintet, J= 6.8 Hz, 1H), 2.23 (d, J = 7.4 Hz, 1H), 1.54 (d, J = 6.6 Hz, 3H).

Intermediate 3 : 2-acetyl-6-fluoro-3-( 3-fluorophenyl)-4H-chromen-4-one

DMSO (5.60 ml, 79.14 mmol) was added to dichloromethane (40 ml), and cooled to – 78°C, followed by oxalyl chloride (3.40 ml, 39.57 mmol). After 10 min., intermediate 2 (6.00 g, 19.78 mmol) in dichloromethane (54 ml) was added dropwise and stirred for 20 min.

Triethylamine (12 ml) was added and stirred for 1 hour. The reaction mixture was quenched with water and extracted with dichloromethane. The organic layer was dried over sodium sulphate and concentrated under reduced pressure. The crude product was purified by column chromatography with ethyl acetate: petroleum ether to afford the title compound as a yellow solid (4.2 g, 71%) which was used as such in the next step.

Intermediate 4: fS)-6-fluoro-3-f3-fluorophenyl)-2-fl-hvdroxyethyl)-4H-chromen-4-one

To intermediate 3 (2.00 g, 6.66 mmol), R-Alpine borane (0.5 M in THF, 20 ml) was added and heated to 60°C for 20 hours. The reaction mixture quenched with 2N HC1, and extracted with ethyl acetate. The organic layer was dried over sodium sulphate and concentrated under reduced pressure. The crude product was purified by column chromatography with ethyl acetate: petroleum ether to afford the title compound as an off-white solid (1.51 g, 75%).

Enantiomeric excess: 94.2%, enriched in the fast eluting isomer (retention time: 8.78 min.) as determined by HPLC on a chiralpak AD-H column.

Intermediate 5: fR)-l-f6-fluoro-3-f3-fluorophenyl)-4-oxo-4H-chromen-2-yl)ethyl 4- chlorobenzoate

To a solution of intermediate 4 (1.45 g, 4.78 mmol) in THF (15 ml), 4-chlorobenzoic acid (0.748 g, 4.78 mmol) and triphenylphosphine (1.88 g, 7.17 mmol) were added and heated to 45°C followed by diisopropylazodicarboxylate (1.4 ml, 7.17 mmol). After 1 hour, the reaction mixture was concentrated and the residue was purified by column chromatography with ethyl acetate: petroleum ether to afford the title compound as an off-white solid (1.81 g, 86%) which was used without purification in the next step. Intermediate 6: fR)-6-fluoro-3-f3-fluorophenyl)-2-fl-hvdroxyethyl)-4H-chromen-4-one

Method A

Intermediate 5 (1.75 g, 3.96 mmol) in methanol (17 ml) was cooled to 10°C, potassium carbonate (0.273 g, 1.98 mmol) was added and stirred for 30 min. The reaction mixture was concentrated, acidified with 2N HCl solution, extracted with ethyl acetate, dried over sodium sulphate and concentrated under reduced pressure. The crude product was purified by column chromatography with ethyl acetate: petroleum ether to afford the title compound as a yellow solid (1.05 g, 87% yield). Enantiomeric excess: 93.6%>, enriched in the late eluting isomer (retention time: 11.12 min.) as determined by HPLC on a chiralpak AD-H column.

Method B

Step-1 [(R)-2-(l-(benzyloxy)ethyl)-6-fluoro-3-(3-fluorophenyl)-4H-chromen-4-one]: To l-(5-fluoro-2-hydroxyphenyl)-2-(3-fluorophenyl)ethanone (11.00 g, 44.31 mmol) in dichloromethane, HATU (33.7 g, 88.63 mmol) and R-(+)2-benzyloxypropionic acid (9.58 g, 53.17 mmol) were added and stirred for 10 min. Triethylamine (66.7 ml, 0.47 mol) was added dropwise and stirred at RT for 24 hours. The reaction mixture was quenched with water, extracted with dichloromethane, dried over sodium sulphate and concentrated under reduced pressure. The crude product was purified by column chromatography with ethyl acetate:

petroleum ether to afford the title compound as a yellow solid (10.5 g, 60%> yield). 1H-NMR (δ ppm, CDCls, 400 MHz): 7.85 (dd, J = 8.1,3 Hz, 1H), 7.58 (dd, J = 9.1, 4.1 Hz, 1H), 7.47-7.39 (m, 1H), 7.39-7.34 (m, 1H), 7.28-7.20 (m, 3H), 7.20-7.14 (m, 2H), 7.16-7.07 (m, 1H), 6.99-6.89 (m, 2H), 4.50-4.31 (m, 3H), 1.56 (d, J = 6.4 Hz, 3H).

Step-2: (R)-2-(l-(benzyloxy)ethyl)-6-fluoro-3-(3-fluorophenyl)-4H-chromen-4-one obtained in Step-1 (10.5 g, 26.69 mmol) in dichloromethane (110 ml) was cooled to 0°C, aluminium chloride (5.35 g, 40.03 mmol) was added portionwise and stirred at RT for 6 hours. The reaction mixture was quenched with 2N HCl solution, extracted with dichloromethane, dried over sodium sulphate and concentrated under reduced pressure. The crude product was purified by column chromatography with ethyl acetate: petroleum ether to afford intermediate 6 a yellow solid (6.1 g, 76% yield). Enantiomeric excess: 97.7%, enriched in the late eluting isomer (retention time: 11.12 min.) as determined by HPLC on a chiralpak AD-H column.

Intermediate 7: 4-bromo-2-fluoro-l-isopropoxybenzene

To a solution of 4-bromo-3-fluorophenol (10 g, 52.35 mmol) in THF (100ml), isopropyl alcohol (4.8 ml, 62.62 mmol) and triphenylphosphine (20.6 g, 78.52 mmol) were added and heated to 45°C followed by diisopropylazodicarboxylate (15.4 ml, 78.52 mmol). The mixture was refluxed for 1 hour, concentrated and the residue was purified by column

chromatography with ethyl acetate: petroleum ether to afford the title compound as a colorless liquid (13.1 g, 99% yield), which was used without purification in the next step.

Intermediate 8: 2-f3-fluoro-4-isopropoxyphenyl)-4,4,5.,5-tetramethyl-l,3i2-dioxaborolane

Potassium acetate (10.52 g, 107.2 mmol) and bis(pinacolato)diboron (15 g, 58.96 mmol) were added to a solution of intermediate 7 (10.52 g, 107.2 mmol) in dioxane (125 ml), and the solution was degassed for 30 min. [l, -Bis(diphenylphosphino)ferrocene]dichloro palladium(II) CH2CI2 (4.4 g, 5.36 mmol) was added under nitrogen atmosphere and heated to 80°C. After 12 hours, the reaction mixture was filtered through celite and concentrated. The crude product was purified by column chromatography with ethyl acetate: petroleum ether to afford the title compound as a yellow oil (13.9g, 99%) which was used without purification in the next step.

Intermediate 9: 3-f3-fluoro-4-isopropoxyphenyl)-lH-pyrazolo[3.,4-dlpyrimidin-4-amine

To a solution of 3-iodo-lH-pyrazolo[3,4-d]pyrimidin-4-amine (11.0 g, 42.14 mmol) in DMF (110 ml), ethanol (55 ml) and water (55 ml), intermediate 8 (23.4 g, 84.28 mmol) and sodium carbonate (13.3 g, 126.42 mmol) were added and degassed for 30 min.

Tetrakis(triphenylphosphine)palladium(0) (2.4 g, 2.10 mmol) was added under nitrogen atmosphere and heated to 80°C. After 12 hours, the reaction mixture was filtered through celite, concentrated and extracted with ethyl acetate. The organic layer was dried over sodium sulphate and concentrated under reduced pressure. The crude product was triturated with diethyl ether, filtered and dried under vacuum to afford the title compound as light brown solid (3.2 g, 26% yield) which is used as such for the next step.

(RS)- 2-fl-f4-amino-3-f3-fluoro-4-isopropoxyphenyl)-lH-pyrazolo[3.,4-(ilpyrimi(iin-l- yl)ethyl)-6-fluoro-3-(3-fluorophenyl)-4H-chromen-4-one

To a solution of intermediate 9 (0.080 g, 0.293 mmol) in DMF (2 ml), potassium carbonate (0.081 g, 0.587 mmol) was added and stirred at RT for 10 min. To this mixture intermediate 1 (0.215 g, 0.587 mmol) was added and stirred for 12 hours. The reaction mixture was diluted with water and extracted with ethyl acetate. The organic layer was dried over sodium sulphate and concentrated under reduced pressure. The crude product was purified by column chromatography with methanol: dichloromethane to afford the title compound as a pale yellow solid (0.045 g). MP: 175-177°C. 1H-NMR (δ ppm, DMSO-D6, 400 MHz): δ 8.20 (s, 1H), 7.85 (dd, J = 81, 3.0 Hz, 1H), 7.48-7.33 (m, 5H), 7.14 (t, J= 8.3 Hz, 1H), 7.02 (m, 2H), 6.90 (m, 1H), 6.10 (q, J = 7.1 Hz, 1H), 5.42 (s, 2H), 4.64 (quintet, J = 6.0 Hz, 1H), 1.99 (d, J = 7.1 Hz, 3H), 1.42 (d, J= 6.1 Hz, 6H).

fS)-2-fl-f4-amino-3-f3-fluoro-4-isopropoxyphenyl)-lH-pyrazolo[3.,4-(ilpyrimi(iin-l- yl)ethyl)-6-fluoro-3-(3-fluorophenyl)-4H-chromen-4-one (“S-isomer”)

To a solution of intermediate 9 (0.134 g, 0.494 mmol) in THF (2.0 ml), intermediate 6 (0.150 g, 0.494 mmol) and triphenylphosphine (0.194 g, 0.741 mml) were added and stirred at RT for 5 min. Diisopropylazodicarboxylate (0.15 ml, 0.749 mmol) was added heated to 45°C. After 2 hours, the reaction mixture was quenched with water and extracted with ethyl acetate. The organic layer was dried over sodium sulphate and concentrated under reduced pressure. The crude product was purified by column chromatography with ethyl acetate : petroleum ether to afford the title compound as an off-white solid (0.049 g, 20 % yield). MP: 139-142°C. Mass: 571.7 (M+). Enantiomeric excess: 89.8% as determined by HPLC on a chiralpak AD-H column, enriched in the fast eluting isomer (retention time = 10.64 min.). fR)-2-fl-f4-amino-3-f3-fluoro-4-isopropoxyphenyl)-lH-pyrazolo[3.,4-(ilpyrimi(iin-l- yl)ethyl)-6-fluoro-3-(3-fluorophenyl)-4H-ehromen-4-one

To a solution of intermediate 8 (0.284 g, 0.989 mmol) in THF (5.0 ml), intermediate 4 (0.250 g, 0.824 mmol) and tris(4-methoxy)phenylphosphine (0.435 g, 1.23 mml) were added and stirred at RT for 5 min. Diisopropylazodicarboxylate (0.25 ml, 1.23 mmol) was added stirred at RT. After 12 hours, the reaction mixture was quenched with water and extracted with ethyl acetate. The organic layer was dried over sodium sulphate and concentrated under reduced pressure. The crude product was purified by column chromatography with ethyl acetate :

petroleum ether to afford the title compound as an off-white solid (0.105 g, 22 % yield). MP: 145-148°C. Mass: 571.7 (M+). Enantiomeric excess: 95.4% as determined by HPLC on a chiralpak AD-H column, enriched in the late eluting isomer (retention time = 14.83 min.).

PATENT

WO 2014006572

http://www.google.com/patents/WO2014006572A1?cl=en

Figure imgf000005_0001B1 IS DESIRED

(S)-2- (l-(4-amino-3-(3-fluoro-4-isopropoxyphenyl)-lH-pyrazolo[3,4-d]pyrimidin-l-yl)ethyl)-6- fluoro-3-(3-fluorophenyl)-4H-chromen-4-one (compound-B l)

Intermediate 11

[119] Intermediate 11: 4-bromo-2-fluoro-l-isopropoxybenzene:To a solution of 4-bromo-2- fluorophenol (lOg, 52.35 mmol) in THF (100ml), isopropyl alcohol (4.8ml, 62.62 mmol) and triphenylphosphine (20.6g, 78.52 mmol) were added and heated to 45 C followed by diisopropylazodicarboxylate (15.4ml, 78 52 mmol). The mixture was refluxed for lh, concentrated and the residue was purified by column chromatography with ethyl acetate: petroleum ether to afford the title compound as a colourless liquid (13. lg, 99%) which was used without purification in the next step. Intermediate 12

[120] Intermediate 12: 2-(3-fluoro-4-isopropoxyphenyl)-4,4,5,5-tetramethyl- 1,3,2- dioxaborolane: Potassium acetate (10.52 g, 107.2 mmol) and bis(pinacolato)diboron (15g, 58.96 mmol) were added to a solution of intermediate 11 (10.52 g, 107.2 mmol) in dioxane (125 ml), and the solution was degassed for 30 min. [1,1 ‘- Bis(diphenylphosphino)ferrocene]dichloro palladium(II).CH2Cl2 (4.4g, 5.36 mmol) was added under nitrogen atmosphere and heated to 80°C. After 12h the reaction mixture was filtered through celite and concentrated. The crude product was purified by column chromatography with ethyl acetate: petroleum ether to afford the title compound as a yellow oil (13.9g, 99%) which was used without purification in the next step.

Intermediate 13

[121] Intermediate 13: 3-(3-fluoro-4-isopropoxyphenyl)-lH-pyrazolo[3,4-d]pyrimidin-4- amine: To a solution of 3-iodo-lH-pyrazolo[3,4-d]pyrimidin-4-amine (11.0 g, 42.14 mmol) in DMF 110 ml), ethanol (55 ml) and water (55 ml), intermediate 12 (23.4 g, 84.28 mmol) and sodium carbonate (13.3 g, 126.42 mmol) were added and degassed for 30 min. Tetrakis(triphenylphosphine)palladium(0) (2.4 g, 2.10 mmol) was added under nitrogen atmosphere and heated to 80°C. After 12h, the reaction mixture was filtered though celite, concentrated and extracted with ethyl acetate. The organic layer was dried over sodium sulphate and concentrated under reduced pressure. The crude product was triturated with diethyl ether, filtered and dried under vacuum to afford the title compound as light brown solid (3.2 g, 26% yield) which is used as such for the next step.

Example Bl

(S)-2-(l-(4-amino-3-(3-fluoro-4-isopropoxyphenyl)-lH-pyrazolo[3,4-d]pyrimidin-l- yl)ethyl)-6-fluoro-3-(3-fluorophenyl)-4H-chromen-4-one

[127] To a solution of intermediate 13 (0.134 g, 0.494 mmol) in THF (2.0 ml), intermediate 5 (0.150 g, 0.494 mmol) and triphenylphosphine (0.194 g, 0.741 mml) were added and stirred at RT for 5 min. Diisopropylazodicarboxylate ( 0.15 ml, 0.749 mmol) was added heated to 45°C. After 2h, the reaction mixture was quenched with with water and extracted with ethyl acetate. The organic layer was dried over sodium sulphate and concentrated under reduced pressure. The crude product was purified by column chromatography with ethyl acetate : petroleum ether to afford the title compound as an off-white solid (0.049 g, 20 %). MP: 139- 142°C. Mass : 571.7 (M H-NMR (δ ppm, CDC13, 400 MHz): 8.24 (s, 1H), 7.85 (dd, J = 8.2,3.1 Hz, 1H), 7.50-7.29 (m, 5H), 7.14 (t, J = 8.4 Hz, 1H), 7.02 (m, 2H), 6.92 (d, J = 8.4 Hz, 1H), 6.11 (q, J = 7.1 Hz, 1H), 5.40 (s, 2H), 4.66 (quintet, J = 6.1 Hz, 1H), 2.00 (d, J = 7.1Hz, 3H), 1.42 (d, J = 6.1 Hz, 6H). Enantiomeric excess: 89.8% as determined by HPLC on a chiralpak AD-H column, enriched in the fast eluting isomer (retention time = 10.64min.).

PATENT

US 2014/0011819 describe the synthesis of TGR-1202 (Example B l)

http://www.google.co.in/patents/US20140011819

Example B1 (S)-2-(1-(4-amino-3-(3-fluoro-4-isopropoxyphenyl)-1H-pyrazolo[3,4-d]pyrimidin-1-yl)ethyl)-6-fluoro-3-(3-fluorophenyl)-4H-chromen-4-one

  • To a solution of intermediate 13 (0.134 g, 0.494 mmol) in THF (2.0 ml), intermediate 5 (0.150 g, 0.494 mmol) and triphenylphosphine (0.194 g, 0.741 mml) were added and stirred at RT for 5 min. Diisopropylazodicarboxylate (0.15 ml, 0.749 mmol) was added heated to 45° C. After 2 h, the reaction mixture was quenched with with water and extracted with ethyl acetate. The organic layer was dried over sodium sulphate and concentrated under reduced pressure. The crude product was purified by column chromatography with ethyl acetate:petroleum ether to afford the title compound as an off-white solid (0.049 g, 20%). MP: 139-142° C. Mass: 571.7 (M+).1H-NMR (δ ppm, CDCl3, 400 MHz): 8.24 (s, 1H), 7.85 (dd, J=8.2, 3.1 Hz, 1H), 7.50-7.29 (m, 5H), 7.14 (t, J=8.4 Hz, 1H), 7.02 (m, 2H), 6.92 (d, J=8.4 Hz, 1H), 6.11 (q, J=7.1 Hz, 1H), 5.40 (s, 2H), 4.66 (quintet, J=6.1 Hz, 1H), 2.00 (d, J=7.1 Hz, 3H), 1.42 (d, J=6.1 Hz, 6H). Enantiomeric excess: 89.8% as determined by HPLC on a chiralpak AD-H column, enriched in the fast eluting isomer (retention time=10.64 min)

4-Methylbenzenesulfonate Salt of Compound B1 (S)-2-(1-(4-amino-3-(3-fluoro-4-isopropoxyphenyl)-1H-pyrazolo[3,4-d]pyrimidin-1-yl)ethyl)-6-fluoro-3-(3-fluorophenyl)-4H-chromen-4-one 4-methylbenzenesulfonate

  • (S)-2-(1-(4-amino-3-(3-fluoro-4-isopropoxyphenyl)-1H-pyrazolo[3,4-d]pyrimidin-1-yl)ethyl)-6-fluoro-3-(3-fluorophenyl)-4H-chromen-4-one 4-methylbenzenesulfonate: To (S)-2-(1-(4-amino-3-(3-fluoro-4-isopropoxyphenyl)-1H-pyrazolo[3,4-d]pyrimidin-1-yl)ethyl)-6-fluoro-3-(3-fluorophenyl)-4H-chromen-4-one (22.7 g, 39.69 mmol) in isopropanol (600 ml), p-toluenesulphonic acid (8.30 g, 43.66 mmol) was added and refluxed for 1 h. The reaction mixture was concentrated, co-distilled with petroleum ether and dried. To the residue water (300 ml) was added and stirred for 30 min. The solid was filtered, washed with petroleum ether and dried under vacuum to afford the title compound as off-white solid (28.2 g, 95%). MP: 138-141° C. 1H-NMR (δ ppm, CDCl3, 400 MHz): 8.11 (s, 1H), 7.85 (dd, J=8.0, 3.0 Hz, 1H), 7.80 (d, J=8.2 Hz, 2H), 7.51 (dd, J=9.3, 4.3 Hz, 1H), 7.45 (dd, J=7.5, 3.1 Hz, 1H), 7.42-7.31 (m, 3H), 7.29 (m, 2H), 7.22 (d, J=8.0 Hz, 2H), 7.16 (t, J=8.3 Hz, 1H), 7.08 (dt, J=8.5, 2.5 Hz, 1H), 6.97 (br s, 1H), 6.88 (br s, 1H), 6.11 (q, J=7.2 Hz, 1H), 4.67 (quintet, J=6.0 Hz, 1H), 2.36 (s, 3H), 2.03 (d, J=7.1 Hz, 3H), 1.43 (d, J=6.0 Hz, 6H). Mass: 572.4 (M++1-PTSA). Enantiomeric excess: 93.4% as determined by HPLC on a chiralpak AD-H column, enriched in the fast eluting isomer (retention time=12.35 min.)

Sulphate Salt of Compound B1 (S)-2-(1-(4-amino-3-(3-fluoro-4-isopropoxyphenyl)-1H-pyrazolo[3,4-d]pyrimidin-1-yl)ethyl)-6-fluoro-3-(3-fluorophenyl)-4H-chromen-4-one sulfate

  • (S)-2-(1-(4-amino-3-(3-fluoro-4-isopropoxyphenyl)-1H-pyrazolo[3,4-d]pyrimidin-1-yl)ethyl)-6-fluoro-3-(3-fluorophenyl)-4H-chromen-4-one sulphate: To (S)-2-(1-(4-amino-3-(3-fluoro-4-isopropoxyphenyl)-1H-pyrazolo[3,4-d]pyrimidin-1-yl)ethyl)-6-fluoro-3-(3-fluorophenyl)-4H-chromen-4-one (15.0 g, 26.24 mmol) in isopropanol (600 ml) was cooled to 0° C. To this Sulphuric acid (2.83 g, 28.86 mmol) was added and stirred at room temperature for 24 h. The reaction mass was filtered and washed with petroleum ether and dried under vacuum. To the solid, water (150 ml) was added and stirred for 30 min. The solid was filtered, washed with petroleum ether and dried under vacuum to afford the title compound as off-white solid (13.5 g, 76%). MP: 125-127° C. 1H-NMR (δ ppm, CDCl3, 400 MHz): 8.11 (s, 1H), 7.85 (dd, J=8.0, 3.0 Hz, 1H), 7.51 (dd, J=9.2, 4.2 Hz, 1H), 7.45-7.31 (m, 3H), 7.29 (m, 1H), 7.15 (t, J=8.3 Hz, 1H), 7.08 (dt, J=8.5, 2.4 Hz, 1H), 6.96 (br s, 1H), 6.88 (br s, 1H), 6.09 (q, J=7.1 Hz, 1H), 4.676 (quintet, J=6.1 Hz, 1H), 2.01 (d, J=7.1 Hz, 3H), 1.42 (d, J=6.1 Hz, 6H). Mass: 572.2 (M++1-H2SO4). Enantiomeric excess: 89.6% as determined by HPLC on a chiralpak AD-H column, enriched in the fast eluting isomer (retention time=12.08 min.)
  • Various other acid addition salts of compound B1 were prepared as provided in Table 1.
  • TABLE 1
    Melting
    Point
    Acid Method of preparation (° C.)
    Hydro- Compound B1 (1 eq.) dissolved in THF, 130-132
    chloric excess HCl/Et2O was added, the clear
    acid solution obtained was evaporated
    completely. The residue obtained was
    washed with water.
    p- Compound B1 (1 eq.) dissolved in 138-141° C.
    Toluene- isopropyl alcohol (IPA), refluxed for
    sulfonic 30 min., acid (1.1 eq.) in IPA was added,
    acid the clear solution obtained was
    evaporated completely. The residue
    obtained was washed with water.
    Benzene- Compound B1 (1 eq.) dissolved in IPA, 170-172
    sulphonic refluxed for 30 min., acid(1.1 eq.) in IPA
    acid was added, the clear solution not
    obtained, the residue was evaporated
    completely and was washed with water.
    Maleic Compound B1 (1 eq.) dissolved in IPA, 107-109
    acid refluxed for 30 min., acid (1.1 eq.) in IPA
    was added, the clear solution not
    obtained, the residue was evaporated
    completely and was washed with water.
    Camphor Compound B1 (1 eq.) dissolved in IPA, 120-121
    sulfonic refluxed for 30 min., acid (1.1 eq.) in IPA
    acid was added, the clear solution not
    obtained, the residue was evaporated
    completely and was washed with water.
    Sulphuric Compound B1 (1 eq.) dissolved in IPA, 125-127
    acid refluxed for 30 min., acid(1.1 eq.) in IPA
    was added, the clear solution obtained
    was evaporated completely. The residue
    obtained was washed with water.

REFERENCES

WO 2014/006572 and U.S. Patent Publication No. 2014/0011819,

http://www.tgtherapeutics.com/O’ConnorTGR202Single%20AgentEHA&Lugano2015.pdf

  • Week in Review, Clinical Status
    Rhizen Pharmaceuticals S.A., La Chaux-de-Fonds, Switzerland TG Therapeutics Inc. (NASDAQ:TGTX), New York, N.Y. Product: TGR-1202 (formerly RP5264) Business: Cancer Molecular target: Phosphoinositide 3-kinase (PI3K) …
  • Week in Review, Clinical Status
    LFB S.A., Les Ulis, France TG Therapeutics Inc. (NASDAQ:TGTX), New York, N.Y. Product: Ublituximab (TGTX-1101, TG-1101, LFB-R603) Business: Cancer Molecular target: CD20 Description: Glycoengineered mAb against CD20 …
  • The Daily Extra, Company News
    TG Therapeutics Inc. (NASDAQ:TGTX) rose $2.65 (23%) to $14.37 after the company said it received an SPA from FDA for the Phase III UNITY-CLL trial of ublituximab (TG-1101) in combination with TGR-1202 to treat chronic …
  • BC Innovations, Targets & Mechanisms
    Now that Nimbus has put IRAK4 on the map for B cell lymphoma, several companies are closing in with their own inhibitors, and they’re all on track for IND-enabling studies this year.
  • Week in Review, Clinical Results
    Rhizen Pharmaceuticals S.A., La Chaux-de-Fonds, Switzerland TG Therapeutics Inc. (NASDAQ:TGTX), New York, N.Y. Product: TGR-1202 (formerly RP5264) Business: Cancer Molecular target: Phosphoinositide 3-kinase (PI3K) …
  • Week in Review, Clinical Results
    LFB S.A., Les Ulis, France TG Therapeutics Inc. (NASDAQ:TGTX), New York, N.Y. Ildong Pharmaceutical Co. Ltd. (KSE:000230), Seoul, South Korea Product: Ublituximab (TGTX-1101, TG-1101, LFB-R603) Business: Cancer …
  • Week in Review, Clinical Status
    Rhizen Pharmaceuticals S.A., La Chaux-de-Fonds, Switzerland TG Therapeutics Inc. (NASDAQ:TGTX), New York, N.Y. Product: TGR-1202 (formerly RP5264) Business: Cancer Molecular target: Phosphoinositide 3-kinase (PI3K) …
  • Week in Review, Deals
    Rhizen Pharmaceuticals S.A., La Chaux-de-Fonds, Switzerland TG Therapeutics Inc. (NASDAQ:TGTX), New York, N.Y. Business: Cancer TG Therapeutics exercised an option under a 2012 deal to license exclusive, worldwide …
Patent Submitted Granted
NOVEL SELECTIVE PI3K DELTA INHIBITORS [US2014011819] 2013-07-02 2014-01-09
Treatment Of Cancers Using PI3 Kinase Isoform Modulators [US2014377258] 2014-05-30 2014-12-25

////////Umbralisib

CC(C)OC1=C(C=C(C=C1)C2=NN(C3=C2C(=NC=N3)N)C(C)C4=C(C(=O)C5=C(O4)C=CC(=C5)F)C6=CC(=CC=C6)F)F
RP 6530, Tenalisib


str1RP 6530

(S)-2-(l-(9H-purin-6-ylamino)propyl)-3-(3-fluorophenyl)-4H-chromen-4-one (Compound A1 is RP 6530).

str1

RP 6530

CID 86291103.png

RP 6530, RP6530, RP-6530

Tenalisib

RP6530-1401, NCI-2015-01804, 124584, NCT02567656

(S)-2-(l-(9H-purin-6-ylamino)propyl)-3-(3-fluorophenyl)-4H-chromen-4-one

3-(3-fluorophenyl)-2-[(1S)-1-(7H-purin-6-ylamino)propyl]chromen-4-one

MW415.4, C23H18FN5O2

CAS 1639417-53-0, 1693773-94-2

RP6530 demonstrated high potency against PI3Kδ (IC50 =24.5 nM) and γ (IC50 = 33.2 nM) enzymes with selectivity over α (>300-fold) and β (>100-fold) isoforms. Cellular potency was confirmed in target-specific assays, namely anti-FcεR1-(EC50=37.8 nM) or fMLP (EC50 = 39.0 nM) induced CD63 expression in human whole blood basophils, LPS induced CD19+ cell proliferation in human whole blood (EC50=250 nM), and LPS induced CD45R+ cell proliferation in mouse whole blood (EC50=101 nM).
A PI3K inhibitor potentially for the treatment of hematologic malignancies.

An inhibitor of phosphoinositide-3 kinase (PI3K) δ/γ isoforms and anti-cellular proliferation agent for treatment of hematol. malignancies

Rhizen Pharmaceuticals is developing RP-6530, a PI3K delta and gamma dual inhibitor, for the potential oral treatment of cancer and inflammation  In November 2013, a phase I trial in patients with hematologic malignancies was initiated in Italy ]\. In September 2015, a phase I/Ib study was initiated in the US, in patients with relapsed and refractory T-cell lymphoma. At that time, the study was expected to complete in December 2016

PATENTS……..WO 11/055215 ,  WO 12/151525.

Inventors

 Inventors Meyyappan Muthuppalaniappan, Srikant Viswanadha, Govindarajulu Babu, Swaroop Kumar V.S. Vakkalanka,
Incozen Therapeutics Pvt. Ltd., Rhizen Pharmaceuticals Sa
  • Antineoplastics; Small molecules
  • Mechanism of Action Phosphatidylinositol 3 kinase delta inhibitors; Phosphatidylinositol 3 kinase gamma inhibitors
  • Phase I Haematological malignancies
  • Preclinical Multiple myeloma
Swaroop K. V. S. Vakkalanka,
COMPANY Rhizen Pharmaceuticals Sa

https://clinicaltrials.gov/ct2/show/NCT02017613

PI3K delta/gamma inhibitor RP6530 An orally active, highly selective, small molecule inhibitor of the delta and gamma isoforms of phosphoinositide-3 kinase (PI3K) with potential immunomodulating and antineoplastic activities. Upon administration, PI3K delta/gamma inhibitor RP6530 inhibits the PI3K delta and gamma isoforms and prevents the activation of the PI3K/AKT-mediated signaling pathway. This may lead to a reduction in cellular proliferation in PI3K delta/gamma-expressing tumor cells. In addition, this agent modulates inflammatory responses through various mechanisms, including the inhibition of both the release of reactive oxygen species (ROS) from neutrophils and tumor necrosis factor (TNF)-alpha activity. Unlike other isoforms of PI3K, the delta and gamma isoforms are overexpressed primarily in hematologic malignancies and in inflammatory and autoimmune diseases. By selectively targeting these isoforms, PI3K signaling in normal, non-neoplastic cells is minimally impacted or not affected at all, which minimizes the side effect profile for this agent. Check for active clinical trials using this agent. (NCI Thesaurus)

Company Rhizen Pharmaceuticals S.A.
Description Dual phosphoinositide 3-kinase (PI3K) delta and gamma inhibitor
Molecular Target Phosphoinositide 3-kinase (PI3K) delta ; Phosphoinositide 3-kinase (PI3K) gamma
Mechanism of Action Phosphoinositide 3-kinase (PI3K) delta inhibitor; Phosphoinositide 3-kinase (PI3K) gamma inhibitor
Therapeutic Modality Small molecule

str1

str1

Dual PI3Kδ/γ Inhibition By RP6530 Induces Apoptosis and Cytotoxicity In B-Lymphoma Cells

RP6530 is a potent and selective dual PI3Kδ/γ inhibitor that inhibited growth of B-cell lymphoma cell lines with a concomitant reduction in the downstream biomarker, pAKT. Additionally, the compound showed cytotoxicity in a panel of lymphoma primary cells. Findings provide a rationale for future clinical trials in B-cell malignancies.

PI3K Dual Inhibitor (RP-6530)


Therapeutic Area Respiratory , Oncology – Liquid Tumors , Rheumatology Molecule Type Small Molecule
Indication Peripheral T-cell lymphoma (PTCL) , Non-Hodgkins Lymphoma , Asthma , Chronic Obstructive Pulmonary Disease (COPD) , Rheumatoid Arthritis
Development Phase Phase I Rt. of Administration Oral

Description

Rhizen is developing dual PI3K gamma/delta inhibitors for liquid tumors and inflammatory conditions.

Mechanism of Action

While alpha and beta isoforms are ubiquitous in their distribution, expression of delta and gamma is restricted to circulating hematogenous cells and endothelial cells. Unlike PI3K-alpha or beta, mice lacking expression of gamma or delta do not show any adverse phenotype indicating that targeting of these specific isoforms would not result in overt toxicity. Dual delta/gamma inhibition is strongly implicated as an intervention strategy in allergic and non-allergic inflammation of the airways and other autoimmune diseases. Scientific evidence for PI3K-delta and gamma involvement in various cellular processes underlying asthma and COPD stems from inhibitor studies and gene-targeting approaches. Also, resistance to conventional therapies such as corticosteroids in several COPD patients has been attributed to an up-regulation of the PI3K delta/gamma pathway. Disruption of PI3K-delta/gamma signalling therefore provides a novel strategy aimed at counteracting the immuno-inflammatory response. Due to the pivotal role played by PI3K-delta and gamma in mediating inflammatory cell functionality such as leukocyte migration and activation, and mast cell degranulation, blocking these isoforms may also be an effective strategy for the treatment of rheumatoid arthritis as well.

Given the established criticality of these isoforms in immune surveillance, inhibitors specifically targeting the delta and gamma isoforms would be expected to attenuate the progression of immune response encountered in airway inflammation and rheumatoid arthritis.

https://i0.wp.com/www.rhizen.com/images/backgrounds/pi3k%20delta%20gamma%20ii.png

Clinical Trials

Rhizen has identified an orally active Lead Molecule, RP-6530, that has an excellent pre-clinical profile. RP-6530 is currently in non-GLP Tox studies and is expected to enter Clinical Development in H2 2013.

In December 2013, Rhizen announced the start of a Phase I clinical trial. The study entitled A Phase-I, Dose Escalation Study to Evaluate Safety and Efficacy of RP6530, a dual PI3K delta /gamma inhibitor, in patients with Relapsed or Refractory Hematologic Malignancies is designed primarily to establish the safety and tolerability of RP6530. Secondary objectives include clinical efficacy assessment and biomarker response to allow dose determination and potential patient stratification in subsequent expansion studies.

Partners by Region

Rhizen’s pipeline consists of internally discovered (with 100% IP ownership) novel small molecule programs aimed at high value markets of Oncology, Immuno-inflammtion and Metabolic Disorders. Rhizen has been successful in securing critical IP space in these areas and efforts are on for further expansion in to several indications. Rhizen seeks partnerships to unlock the potential of these valuable assets for further development from global pharmaceutical partners. At present global rights on all programs are available and Rhizen is flexible to consider suitable business models for licensing/collaboration.

In 2012, Rhizen announced a joint venture collaboration with TG Therapeutics for global development and commercialization of Rhizen’s Novel Selective PI3K Kinase Inhibitors. The selected lead RP5264 (hereafter, to be developed as TGR-1202) is an orally available, small molecule, PI3K specific inhibitor currently being positioned for the treatment of haematological malignancies.

PATENT

WO2014195888, DUAL SELECTIVE PI3 DELTA AND GAMMA KINASE INHIBITORS

https://patentscope.wipo.int/search/en/detail.jsf?docId=WO2014195888Intermediates

Intermediate 1: 3-(3-fluorophenyl)-2-(l-hydroxypropyl)-4H-chromen-4-one: To a solution of 2-(l-bromopropyl)-3-(3-fluorophenyl)-4H-chromen-4-one1 (8.80 g, 24.36 mmol ) in DMSO (85 ml), n-butanol (5 ml) was added and heated to 120° C for 3h. The reaction mixture was cooled to room temperature (RT), quenched with water and extracted with ethyl acetate. The organic layer was dried over sodium sulphate and concentrated under reduced pressure. The crude product was purified by column chromatography with ethyl acetate: petroleum ether to afford the title compound as a yellow solid (2.10 g, 29 %) which was used without further purification in next step.

Intermediate 2: 3-(3-fluorophenyl)-2-propionyl-4H-chromen-4-one: DMSO (1.90 ml, 26.82 mmol) was added to dichloromethane (70 ml) and cooled to -78°C. Oxalyl chloride (1.14 ml, 13.41 mmol) was then added. After 10 minutes, intermediate 1 (2.00 g, 6.70 mmol) in dichloromethane (20 ml) was added dropwise and stirred for 20 min. Triethylamine (7 ml) was added and stirred for lh. The reaction mixture was quenched with water and extracted with dichloromethane. The organic layer was dried over sodium sulphate and concentrated under reduced pressure. The crude product was purified by column chromatography with ethyl acetate: petroleum ether to afford the title compound as a yellow liquid (1.20 g, 60%) which was used as such in next step.

Intermediate 3: (+)/(-)-3-(3-fluorophenyl)-2-(l-hydroxypropyl)-4H-chromen-4-one :

To a solution of intermediate 2 (0.600 g, 2.02 mmol) in DMF (7.65 ml) under nitrogen purging, formic acid : trietylamine 5 : 2 azeotrope (1.80 ml) was added followed by [(S,S)tethTsDpenRuCl] (3.0 mg). The reaction mixture was heated at 80°C for 1.5 hours under continuous nitrogen purging. The reaction mixture was quenched with water, extected with ethyl acetate, dried over sodium sulphate and concentrated. The crude product was purified by column chromatography with ethyl acetate: petroleum ether to afford the title compound as a yellow solid (0.450 g, 74%). Mass: 299.0 (M+).

Enantiomeric excess: 78%, enriched in the late eluting isomer (retention time: 9.72 min.) as determined by HPLC on a chiralpak AD-H column.

Intermediate 4: (+)/(-)-3-(3-fluorophenyl)-2-(l-hydroxypropyl)-4H-chromen-4-one :

The title compound was obtained as yellow solid (0.500 g, 83%) by using a procedure similar to the one described for intermediate 3, using intermediate 2 (0.600 g, 2.02 mmol), DMF (7.65 ml), formic acid : trietylamine 5 : 2 azeotrope (1.80 ml) and [(R,R)tethTsDpenRuCl] (3.0 mg). Mass: 298.9 (M+). Enantiomeric excess: 74.8%, enriched in the fast eluting isomer (retention time: 8.52 min.) as determined by HPLC on a chiralpak AD-H column.

Intermediate 5: (R)-3-(3-fluorophenyl)-2-(l-hydroxypropyl)-4H-chromen-4-one:

Step 1 : (R)-2-(l-(benzyloxy)propyl)-3-(3-fluorophenyl)-4H-chromen-4-one: To 2-(3-fluorophenyl)-l-(2-hydroxyphenyl)ethanone (2.15 g, 9.36 mmol ), in dichloromethane ( 20 ml), HATU (4.27 g, 11.23 mmol), R-(+)2-benzyloxybutyric acid (2.00 g, 10.29 mmol) were added and stirred for lOmin, then triethylamine (14.0 ml, 101.1 mmol) was added dropwise and stirred at RT for 24h. The reaction mixture was quenched with water, extracted with dichloromethane, dried over sodium sulphate and concentrated under reduced pressure. The crude product was purified by column chromatography with ethyl acetate: petroleum ether to afford the title compound as yellow solid (1.65 g, 45%). JH-NMR (δ ppm, CDC13, 400 MHz): 8.24 (dd, / = 7.9,1.5 Hz, 1H), 7.74 (dt, / = 7.1,1.7 Hz, 1H), 7.58 (dd, / = 8.3,0.4 Hz, 1H), 7.44-7.06 (m, 10H), 4.51 (d, / = 7.8 Hz, 1H), 4.34 (d, / = 7.8 Hz, 1H), 4.25 (dd, / = 7.8,6.2 Hz, 1H), 2.17-1.90 (m, 2H), 0.95 (t, / = 7.5 Hz, 3H). Mass: 389.0 (M+).

Step 2: (R)-3-(3-fluorophenyl)-2-(l-hydroxypropyl)-4H-chromen-4-one : To (R)-2-(l-(benzyloxy)propyl)-3-(3-fluorophenyl)-4H-chromen-4-one (1.50 g, 3.86 mmol) in dichloromethane (15 ml) cooled to 0°C and aluminium chloride (1.00 g, 7.72 mmol) was added portion wise and stirred at RT for 6h. The reaction mixture was quenched with 2N HC1 solution, extracted with dichloromethane, dried over sodium sulphate and concentrated under reduced pressure. The crude product was purified by column chromatography with ethyl acetate: petroleum ether to afford the title compound as yellow solid (0.552 g, 48%).‘ JH-NMR (δ ppm, CDC13, 400 MHz): 8.24 (dd, / = 8.0,1.6 Hz, 1H), 7.72 (m, , 1H), 7.52 (dd, / = 8.4,0.5 Hz, 1H), 7.44 (m, 2H), 7.12-7.01(m,3H), 4.49 (t, / = 7.0 Hz, 1H), 1.94 (m, 2H), 0.93 (t, / = 7.5 Hz, 3H). Mass: (299.0(M+). Purity: 96.93%.

25[a] D -14.73 (c = 1, CHCI3). Enantiomeric excess: 85.92%, enriched in the fast eluting isomer (retention time: 8.57 min.) as determined by HPLC on a chiralpak AS-3R column.

Compound A

(RS)- 2-(l-(9H-purin-6-ylamino)propyl)-3-(3-fluorophenyl)-4H-chromen-4-one

To a solution of intermediate 1 (2.50 g, 8.41 mmol) in THF (25 ml), tert-butyl 9-trityl-9H-purin-6-ylcarbamate (4.81 g, 10.09 mmol) and triphenylphosphine (3.31 g, 12.62 mmol) were added and stirred at RT for 5 min. Diisopropylazodicarboxylate (2.5 ml, 12.62 mmol) was added and stirred at RT for 2h. The reaction mixture was concentrated and column chromatographed with ethyl acetate : petroleum ether to afford a yellow coloured intermediate. To the intermediate, dichloromethane (65 ml) and trifluoroacetic acid (7.9 ml) were added and the resulting mixture was stirred at RT for 12 h. The reaction mixture was then basified with aqueous sodium bicarbonate solution, extracted with dichloromethane and dried over sodium sulphate. The crude product was purified by column chromatography with methanol: dichloromethane to afford the title compound as pale-brown solid (1.05 g, 30 %). MP: 148-150°C. Mass: 415.6 (M+).

Compound Al

(S)-2-(l-(9H-purin-6-ylamino)propyl)-3-(3-fluorophenyl)-4H-chromen-4-one

Method A: To a solution of intermediate 3 (0.250 g, 0.838 mmol) in THF (5ml), tert-butyl 9-trityl-9H-purin-6-ylcarbamate (0.479 g, 1.00 mmol) and triphenylphosphine (0.329 g, 1.25 mmol) were added and the resulting mixture was stirred at RT for 5 min. Diisopropylazodicarboxylate (0.25 ml, 1.25 mmol) was then added and stirred at RT for 12 h. The reaction mixture was concentrated and column chromatographed with ethyl acetate: pet.ether to afford the yellow coloured intermediate. To the intermediate in dichloromethane (6 ml), trifluoroacetic acid (1.2 ml) was added stirred at RT for 12 h. The reaction mixture was basified with aqueous sodium bicarbonate solution, extracted with dichloromethane and dried over sodium sulphate. The crude product was purified by column chromatography with methanol: dichloromethane to afford the title compound as an off-white solid (0.015 g, 4 %). MP: 137-140°C. JH-NMR (δ ppm, DMSO- , 400 MHz): 12.94 (s, 1H), 8.12-8.10 (m, 4H), 7.84-7.80 (m, 1H), 7.61 (d, / = 8.3 Hz, 1H), 7.50-7.41 (m, 2H), 7.28-7.18 (m, 3H), 5.20-5.06 (m, 1H), 2.10-1.90 (m, 2H), 0.84 (t, / = 3.7 Hz, 3H). Enantiomeric excess: 77.4% as determined by HPLC on a chiralpak AD-H column, enriched in the fast eluting isomer (retention time = 7.90 min.).

Method B : To a solution of intermediate 5 (2.60 g, 8.68 mmol) in THF (52 ml), tert-butyl 9-trityl-9H-purin-6-ylcarbamate (4.96 g, 10.42 mmol) and triphenylphosphine (2.76 g, 13.03 mmol) were added and the resulting mixture was stirred at RT for 5 min. Dusopropylazodicarboxylate (0.25 ml, 1.25 mmol) was then added and stirred at RT for 12 h. The reaction mixture was concentrated and column chromatographed with ethyl acetate: petroleum ether to afford the yellow coloured intermediate. To the intermediate in dichloromethane (55 ml), trifluoroacetic acid (14.2 ml) was added and stirred at RT for 12 h. The reaction mixture was basified with aqueous sodium bicarbonate solution, extracted with dichloromethane and dried over sodium sulphate. The crude product was purified by column chromatography with methanol: dichloromethane to afford the title compound as pale-yellow solid (1.00 g, 27 %). MP: 168-170°C. Mass: 416.5(M++1) Enantiomeric excess: 86.5% as determined by HPLC on a chiralpak AD-H column, enriched in the fast eluting isomer (retention time = 7.90 min.).

Method C : The title compound was separated by preparative SFC conditions from Compound A (1.090 g) on a CHIRALPAK AY-H column (250 x 30 mm; 5μπι) using methanol : C(¾ (35:65) as the mobile phase at a flow rate of 80 g / min. Off-white solid (0.378 g). e.e. 100%. Rt: 2.37 min. Mass: 416.1(M++1). MP: 149-152°C.

PATENT
WO 2011055215

Scheme 1A

CAUTION        ethyl compd below, NOT THE PRODUCT

Example 47

(S)-2-(l-(9H-purin-6-yIamino) ethyl)-3-(3-fluorophenyl)-4H-chromen-4-one

[428] To a solution of intermediate 65 (2.0g, 8.68 mmoles) in dichloromethane (20ml), triethylamine (3.6ml, 26.06 mmoles) was added followed by N-Boc-Alanine (1.97g, 10.42 mmoles). To this mixture HATU (6.6g, 17.37 mmoles) was added and stirred at RT for 12h. The reaction mixture was quenched by the addition of water and extracted with dichloromethane. The organic layer was dried over sodium sulphate and concentrated under reduced pressure. The crude product was purified by column chromatography with ethyl acetate: petroleum ether to afford the isoflavone intermediate (1.70g). To a solution of this intermediate (1.7g) in dichloromethane (20ml), trifluoroacetic acid (3 ml) was added and stirred at RT for 2h. The reaction mixture was concentrated, basified with sodium bicarbonate solution, extracted with ethyl acetate. The organic layer was dried over sodium sulphate and concentrated under reduced pressure to afford the amine intermediate (0.641 g). To a solution of this amine intermediate (0.30g, 1.05 mmoles) in tert-butanol (6ml), N, N- diisopropylethylamine (0.36ml, 2.17 mmoles) and 6-bromopurine (0.168g, 0.847 mmoles) were added and refluxed for 24h. The reaction mixture was concentrated, diluted with water, extracted with ethyl acetate. The organic layer was dried over sodium sulphate and concentrated under reduced pressure. The crude product was purified by column chromatography with methanol: ethyl acetate to afford the title compound as off-white solid (0.041g, 10% yield). MP: 135-138 °C. Ή-NMR (δ ppm, DMSO-D6, 400 MHz): δ 12.95(s,lH), 8.15(t, / = 6.8Hz, 1H), 8.11(s, 1H), 8.08(s, 1H), 8.03(d, J = 7.8 Hz, 1H), 7.81(t ,J = 7.3Hz, 1H), 7.60 (d, J = 8.3Hz, 1H), 7.49 (t, J = 7.3Hz, 2H), 7.25(m,3H), 5.19(br m, 1H), 1.56(d, J = 6.9Hz,3H). Mass: 402.18(M+ +1).

PATENT
WO 2012151525

Scheme 1

Base

This scheme provides a synthetic route for the preparation of compound of formula wherein all the variables are as described herein in above

15 14 10 12 12a

 CONFERENCE PROCEEDINGS

Abstract 2704: RP6530, a dual PI3K δ/γ inhibitor, potentiates ruxolitinib activity in the JAK2-V617F mutant erythroleukemia cell lines

  1. Swaroop Vakkalanka1,
  2. Seeta Nyayapathy2, and
  3. Srikant Viswanadha2

Author Affiliations

  1. 1Rhizen Pharmaceuticals SA, Fritz-Courvoisier 40, Switzerland;
  2. 2Incozen Therapeutics Pvt. Ltd., Hyderabad, India.
Proceedings: AACR 106th Annual Meeting 2015; April 18-22, 2015; Philadelphia, PA

Abstract

Background: Myelofibrosis (MF) represents a life-threatening neoplasm that manifests particularly in the elderly population and is characterized by bone marrow fibrosis and extramedullary hematopoeisis. While ruxolitinib, a JAK1/2 inhibitor, has recently been approved by the USFDA for its disease modifying potential in MF patients, it is still not considered as a curative option. Targeting another kinase such as PI3K, downstream of JAK, could therefore be a more efficient way of treating myelofibrotic neoplasms. RP6530 is a novel, potent, and selective PI3K δ/γ inhibitor that demonstrated high potency against PI3Kδ (IC50 = 25 nM) and γ (IC50 = 33 nM) enzymes with selectivity over α (>300-fold) and β (>100-fold) isoforms. The objective of this study was to evaluate the effect of a combination of ruxolitinib and RP6530 in the JAK2-V617F mutant Human Erythroleukemia (HEL) cell line.

Methods: Passive resistance was conferred by incubating HEL cells with increasing concentrations of ruxolitinib over an 8-10-week period. Endogenous JAK2, PI3Kδ, PI3Kδ, and pAKT were estimated by Western Blotting. RP6530, ruxolitinib, and the combination of RP6530 + Ruxolitinib were tested for their effect on viability and apoptosis. Cell viability was assessed by a MTT assay. Induction of apoptosis was analyzed by Annexin V/PI staining.

Results: Resistance to ruxolitinib was confirmed by a right-ward shift in EC50 of ruxolitinib in a HEL cell proliferation assay (0.82 μM Vs. 12.2 μM). Endogeous pAKT expression was 3.7-fold higher in HEL-RR compared to HEL-RS cells indicating activation of the AKT signaling pathway. While single-agent activity of RP6530 was modest (33-46% inhibition @ 10 μM) in both HEL-RS and HEL-RR cells, addition of 10 μM RP6530 to ruxolitinib was synergistic resulting in a near-complete inhibition of proliferation (>90% for HEL-RS and >70% for HEL-RR). While the order of addition did not affect the potency of RP6530, addition of 5 μM RP6530, 4 h prior to the addition of ruxolitinib resulted in a significant reduction in EC50 of ruxolitinib (5.8 μM) in HEL-RR cells. On lines with cell proliferation data, incubation of 10 μM RP6530 with ruxolitinib for 72 h increased the percent of apoptotic cells (55% in HEL-RS and 37% in HEL-RR) compared to either agent alone (16-27% in HEL-RS and 17-21% in HEL-RR).

Conclusions: Ruxolitinib resistance in the V617F JAK-2 mutant HEL cells is accompanied by an increase in pAKT expression. Inhibition of pAKT via the addition of RP6530, a dual PI3K δ/γ inhibitor, resulted in a reversal of ruxolitinib resistance. Complementary activity was also observed in HEL-RS cells indicating that a combination of ruxolitinib and RP6530 could have a positive bearing on the clinical outcome in MF patients.

Citation Format: Swaroop Vakkalanka, Seeta Nyayapathy, Srikant Viswanadha. RP6530, a dual PI3K δ/γ inhibitor, potentiates ruxolitinib activity in the JAK2-V617F mutant erythroleukemia cell lines. [abstract]. In: Proceedings of the 106th Annual Meeting of the American Association for Cancer Research; 2015 Apr 18-22; Philadelphia, PA. Philadelphia (PA): AACR; Cancer Res 2015;75(15 Suppl):Abstract nr 2704. doi:10.1158/1538-7445.AM2015-2704

REFERENCES
December 2014, data were presented at the 56th ASH Meeting in San Francisco, CA.
April 2015, preclinical data were presented at the 106th AACR Meeting in Philadelphia, PA. RP-6530 had GI50 values of 17,028 and 22,014 nM, respectively
December 2013, preclinical data were presented at the 55th ASH Meeting in New Orleans, LA.
June 2013, preclinical data were presented at the 18th Annual EHA Congress in Stockholm, Sweden. RP-6530 inhibited PI3K delta and gamma isoforms with IC50 values of 24.5 and 33.2 nM, respectively.
  • 01 Sep 2015 Phase-I clinical trials in Haematological malignancies (Second-line therapy or greater) in USA (PO) (NCT02567656)
  • 18 Nov 2014 Preclinical trials in Multiple myeloma in Switzerland (PO) prior to November 2014
  • 18 Nov 2014 Early research in Multiple myeloma in Switzerland (PO) prior to November 2014
WO2011055215A2 Nov 3, 2010 May 12, 2011 Incozen Therapeutics Pvt. Ltd. Novel kinase modulators
WO2012151525A1 May 4, 2012 Nov 8, 2012 Rhizen Pharmaceuticals Sa Novel compounds as modulators of protein kinases
WO2013164801A1 May 3, 2013 Nov 7, 2013 Rhizen Pharmaceuticals Sa Process for preparation of optically pure and optionally substituted 2- (1 -hydroxy- alkyl) – chromen – 4 – one derivatives and their use in preparing pharmaceuticals
US20110118257 May 19, 2011 Rhizen Pharmaceuticals Sa Novel kinase modulators
US20120289496 May 4, 2012 Nov 15, 2012 Rhizen Pharmaceuticals Sa Novel compounds as modulators of protein kinases
WO 2014195888
WO 2011055215

WO2015175966

WO2015051252

  • BC Innovations, Therapeutics
    Indication Target/marker/pathway Summary Licensing status Publication and contact information Cardiovascular disease Intimal hyperplasia Phosphoinositide 3-kinase-g (PI3Kg) Rodent studies suggest inhibiting …
  • BC Innovations, Targets & Mechanisms
    Targets & Mechanisms: PI3K inhibition: solid immunotherapy Table 1. A peek at PI3K inhibitors. According to a study in Nature by Ali et al., inhibition of phosphoinositide 3-kinase-d (PI3Kd) or the PI3K catalytic …
  • Week in Review, Clinical Status
    Rhizen Pharmaceuticals S.A., La Chaux-de-Fonds, Switzerland Product: RP6530 Business: Cancer Molecular target: Phosphoinositide 3-kinase (PI3K) delta; Phosphoinositide 3-kinase (PI3K) gamma Description: Dual …
  • Week in Review, Preclinical Results
    Rhizen Pharmaceuticals S.A., La Chaux-de-Fonds, Switzerland Product: RP6530 Business: Cancer Indication: Treat B cell lymphoma In vitro, 2-7 M RP6530 led to a &gt;50% dose-dependent inhibition in growth of immortalized …

/////

c1cccc4c1C(/C(=C(/[C@H](CC)Nc3c2c(ncn2)ncn3)O4)c5cc(ccc5)F)=O

CCC(C1=C(C(=O)C2=CC=CC=C2O1)C3=CC(=CC=C3)F)NC4=NC=NC5=C4NC=N5

Identification of Medicinal Products Standards will apply in six Months


DR ANTHONY MELVIN CRASTO Ph.D's avatarDRUG REGULATORY AFFAIRS INTERNATIONAL

The pharma sector must comply with IDMP standards in the EU region starting July 2016. This provides regulators with the means of easily comparing product data across regions and with different manufacturers.

http://www.gmp-compliance.org/enews_05125_Identification-of-Medicinal-Products-Standards-will-apply-in-six-Months_15179,15237,Z-QAMPP_n.html

Over the last couple of years the European Health Authorities in conjunction with the International Standards Organization (ISO) have been developing a set of global data standards referred to as Identification of Medicinal Products (IDMP).

The Identification of Medicinal Products (IDMP) standards were developed in response to a worldwide demand for internally harmonized specifications for medicinal products. The EU is the first to implement these standards, and the other ICH regions will follow. The pharma sector must comply with IDMP standards in the EU region starting July 2016. Following the EU, the other ICH (International Conference on Harmonization of Technical Requirements for Registration of Pharmaceuticals for Human Use) countries will then begin their own adoption processes…

View original post 124 more words

Tazobactam


Tazobactam.svgTazobactam.png


Tazobactam; Tazobactam acid; 89786-04-9; Tazobactamum; YTR-830H; CL-298741

(2S,3S,5R)-3-methyl-4,4,7-trioxo-3-(triazol-1-ylmethyl)-4$l^{6}-thia-1-azabicyclo[3.2.0]heptane-2-carboxylic acid

CAS 89785-84-2 SODIUM SALT

TAIHO Innovator

Molecular Formula: C10H12N4O5S
Molecular Weight: 300.29108 g/mol

Tazobactam is a beta Lactamase Inhibitor. The mechanism of action of tazobactam is as a beta Lactamase Inhibitor.

Tazobactam is a penicillanic acid sulfone derivative and beta-lactamase inhibitor with antibacterial activity. Tazobactam contains a beta-lactam ring and irreversibly binds to beta-lactamase at or near its active site. This protects other beta-lactam antibiotics from beta-lactamase catalysis. This drug is used in conjunction with beta-lactamase susceptible penicillins to treat infections caused by beta-lactamase producing organisms.

Tazobactam is a pharmaceutical drug that inhibits the action of bacterial β-lactamases, especially those belonging to the SHV-1 and TEM groups. It is commonly used as its sodium salt, tazobactam sodium.

Tazobactam is combined with the extended spectrum β-lactam antibiotic piperacillin in the drug piperacillin/tazobactam, one of the preferred antibiotic treatments for nosocomial pneumonia caused by Pseudomonas aeruginosa.[citation needed] Tazobactam broadens the spectrum of piperacillin by making it effective against organisms that express β-lactamase and would normally degrade piperacillin.[1]

Tazobactam is a heavily modified penicillin and a sulfone.


PAPER

doi:10.1021/jm00391a032

http://pubs.acs.org/doi/pdf/10.1021/jm00391a032

Synthesis 

PATENT

CN 104031065

http://www.google.com/patents/CN104031065A?cl=zh

[2S- (2 α, 2 β, 5 α)] -3- methyl _7_ oxo _3_ (1Η-1,2,3_ triazol-1-ylmethyl) -4- thia-1-azabicyclo – [3,2, O] – heptane-2-carboxylic acid 4,4-dioxide.

The structural formula:

Figure CN104031065AD00041

The first from 6-aminopenicillanic acid (6-APA) prepared by starting from the Hall TW et al., Its structure is to add a triazole ring on the basis of sulbactam to improve the effect of inhibiting the enzyme, which is currently lactam best clinical results β_ inhibitor, with high stability, low activity, low toxicity, inhibiting activity and other characteristics. 1992, tazobactam combination drug tazobactam / piperacillin (1: 8) for the first time in France the market, used to treat a variety of bacterial infections.

The literature related to the different synthesis Tazobactam triazole ring according to the introduction, there are two main ways of preparation methods: the azide cycloaddition synthetic triazole five-membered ring and the side chains directly added triazole ring .

Preparation Method One: the azide cycloaddition method, as shown below:

Figure CN104031065AD00051

The azide cycloaddition preparation method, which is penicillanic acid diphenylmethyl ester sulfoxide as raw material, open-loop, chloride, azide, oxidation, alkyne cycloaddition, deprotection steps to obtain cilostazol Batan, although each step is quite simple and easy for industrial production, at present most manufacturers use this route, but its route is longer, and there is the azide reaction byproducts generated a large number of six-membered ring, the total yield compared low.

Preparation Method two: direct plus side chains triazole ring

Direct plus side chains triazole ring Preparation mainly disulfide nucleophilic ring was open and IH-1,2,3- triazole occurred directly in acetic acid in the presence of mercury or mercury oxide substituted rings (US4898939) or directly with the IH-1,2,3- triazole silver salt catalyzed reaction of iodine (Synthesis, 2005,3,442-446), as shown below:

Figure CN104031065AD00052

And the use of methyl chloride in an alkaline environment and iodine catalyzed substitution to generate the target product (CN200810238479 with 1H-1,2,3- triazole; Shanghai Second Medical University, Shanghai 2009/20 (5): 388- 391), as shown below:

Figure CN104031065AD00061

Direct plus side chains triazole ring preparation method because of its short synthetic route, avoiding the risk of high temperature and pressure addition is currently a hot tazobactam drug synthesis research. Since the compound (4) the sulfur atom lone pair of electrons more of a halogen atom (Cl, Br,

I) have a role to leave, under alkaline action by 1H-1,2,3- triazole nucleophilic attack IH ions generated carbocations prone to rearrangement to form a six-membered ring by-products higher probability, if the sulfur atom is oxidized to a sulfone, a sulfur atom, provided no lone pair of electrons, although able to increase its stability, but at the same time a halogen atom (Cl, Br, I) leaving passivation effect, such that the nucleophilic replace hardly occurs while using the expensive raw mercury and silver salts of heavy metals, higher costs, greater environmental pollution, which greatly restricted the industrial scale production.

Synthetic route of the present invention are as follows:

Figure CN104031065AD00071

Example 1: Preparation of 3-methyl – [2-oxo-4- (2-benzothiazolyl dithio) -1-azetidinyl] -3-butene diphenylmethyl ester (Compound 3) Preparation of

In penicillanic acid diphenylmethyl ester sulfoxide (compound 2) as a raw material, according to the literature (Synthesis, 2005,3,442-446) preparation, to give a pale yellow crystalline solid from acetone powder at a yield of 95%.

[0019] Examples of 2: 2 β- bromomethyl -2 α- methyl – penicillanic acid diphenylmethyl ester (Compound 4) Preparation of the solid obtained in Example I (Compound 3) 26g (0.05mol) dissolved in 300mL of methylene chloride, cooled to 0 ° C

The following is added 33.5g (0.075mol) of anhydrous copper bromide, after increases in (T5 ° C the reaction was stirred 10-ΐ2 hours, TLC sample testing of raw materials point disappears, and the filter cake was rinsed with 50mL methylene burn, The filtrate was respectively 200mL water, 200mL saturated sodium bicarbonate, 200mL water washing, containing 2β- bromomethyl -2α- methyl – penicillanic acid diphenylmethyl ester (compound 4) in methylene chloride was used directly in the next step reaction.

Examples 3 [0020]: 2 β – bromomethyl -2 α – methyl – penicillanate _1 β _ oxide diphenylmethyl ester (Compound 5) Preparation of

Of Example 2 was 2 β – bromomethyl -2 α – methyl – penicillanic acid diphenylmethyl ester (Compound 4) in dichloromethane was added 30mL of methanol, cooled to -5 ° C or less, dropwise 30mL50 % hydrogen peroxide / sodium tungstate mixture for about 30 minutes after the dripping, and the temperature at (T5 ° C incubation for 4 hours, then heated to 1 (T15 ° C incubation for 4 flying hours, TLC sample testing of raw materials (Compound 4) disappear , was added 200mL 7jC, stirred for five minutes, standing layer, the liquid layer was then washed with dichloromethane material 200mL 5% aqueous sodium bicarbonate to give comprising 2β – bromomethyl -2 α – methyl – di penicillanate phenylmethyl ester -1 β – oxide (Compound 5) in methylene chloride was used directly in the next reaction.

[0021] Example 4: 2 @ – (! 1 1-1,2,3- triazole group) -20- methyl – penicillanic acid diphenylmethyl ester 1 @ – oxide (compound 6) Preparation

Of Example 3 was 2 β – bromomethyl -2 α – methyl – penicillanic acid diphenylmethyl ester -1 β – oxide (Compound 5) in dichloromethane was added 60mL methanol, 30mL water and 10.35g (0.15mol) 1H-1,2,3- triazole, cooled to below 5 ° C, was added 26g anion resin, temperature 5 ~ 10 ° C and stirred overnight (more than 24 hours), samples of raw materials by TLC (Compound 5) disappears, filtered, and the filtrate was added 200mL of water, standing layered material liquid dichloromethane layer was added anhydrous magnesium sulfate and activated carbon decolorization dehydration process, concentrated and dried under reduced pressure, the residue was added 60mL of methanol was dissolved by heating, stirring slowly cooled to (T5 ° C crystallization, precipitation continued until most of the solids after cooling to below -10 ° C for about 4 hours, filtered, and the cake was rinsed with cold methanol and vacuum dried to give a white solid (compound 6) Hg, yield 82% (Compound 3 by meter), mp: V; ESI (m / z):. 450 ,; IHNMR (CDl3) Examples 5: 2β- (1Η-1,2, 3- triazole-yl) -2α- methyl – penicillanic acid diphenylmethyl ester 1,1-dioxide (Compound 7) Preparation of

The 9g (0.02πιο1) 2β – (1H-1,2,3- triazole group) _2 α – methyl – penicillanic acid diphenylmethyl ester 1-oxide (compound 6) was dissolved in 225mL dichloro methane, adding 45mL glacial acetic acid, cooled to below 0 ° C, was added in portions

3.8g (0.024mol) of potassium permanganate. After the addition was completed in 5 ~ 10 ° C incubated overnight (more than 16 hours), sampled by HPLC completion of the reaction, insolubles were removed by filtration, the filtrate was added to 200mL water, stirred for five minutes, allowed to stand The layers were separated and then washed with 200mL saturated aqueous sodium bicarbonate, methylene chloride stock solution layer was dehydrated by adding anhydrous magnesium sulfate and decolorizing charcoal treatment, the remaining concentrated under reduced pressure to about 50mL volume, slowly with stirring to a cooled (TC hereinafter Crystallization 2 hours, filtered, rinsed with a small amount of methylene chloride, dried in vacuo to give a white solid (Compound 7) 8.85g, yield 95%, mp: 201-206 ° C; ESI (m / z): 466, .; IHNMR (CDl3) Example 6: Preparation of tazobactam he (Compound I),

The 1g (0.021mol) 2 β – (1H-1,2,3- triazole group) _2 α – methyl – penicillanate 1,1-diphenyl ester (compound 7) was dissolved in 10mL m-cresol at 50 ~ 55 ° C incubated for 2 hours, cooled to O ~ 5 ° C, was added 200mL of methyl isobutyl ketone, extracted twice with 10mL saturated sodium bicarbonate solution, the combined aqueous layers were dried 10mL ethyl acetate extract miscellaneous twice, and the aqueous layer was added active carbon filtration, and the filtrate was cooled to O ~ 5 ° C, dropping 6mol / L hydrochloric acid to precipitate a solid no longer far, filtered cake was washed with cold water and dried under vacuum to give a white solid tazobactam 5.9g, yield 92%, mp: 136-1380C; ESI (m / z): 300; IHNMR (CDl3) ο


PATENT


WO 2014037893

http://www.google.com/patents/WO2014037893A1?cl=en

improved process for the preparation of Tazobactam of formula (I).

Figure imgf000003_0001

(I)

Tazobactam is chemically known as 2a-methyl-2 -(l,2,3-triazol-l-yl)- methylpenam-3a-carboxylate- 1,1 -dioxide and has a very low antibacterial activity. On the other hand, it exhibits a beta-lactamase inhibitory activity when irreversibly bonded to beta-lactamases produced by microorganisms. For this reason, Tazobactam may be used in combination with known antibiotics prone to be inactivated by beta-lactamases to allow them to exhibit their inherent antibacterial activity against beta-lactamase producing microorganisms. Tazobactam as a product is disclosed in US Patent No. 4,562,073.

Considering the importance of Tazobactam there are several literatures available which disclose various processes for the preparation of Tazobactam, some of which are described below.

US patent No. 4,562,073 provides Tazobactam of formula (I) and its derivatives. This patent also describes a process for their preparation as shown in Scheme – 1.

Figure imgf000004_0001

(I )

Scheme – 1

wherein R is hydrogen or trialkylsilyl; R is hydrogen, trialkylsilyl or COOM wherein M is hydrogen, C1-18 alkyl, C2-7 alkoxymethyl, etc., R has the same meaning as M and R represents carboxyl protecting group.

US patents 4,891,369 and 4,933,444 disclose an approach, which involves the preparation of 2a-methyl-2 -triazol lmethylpenam derivative of formula (V)

Figure imgf000004_0002

(V)

wherein R is a carboxy protecting group, by treatment of a β-halomethyl penam derivative of formula (IV), wherein X is chlorine or bromine and R is a carboxy protecting group, with 1,2,3-triazole.

Figure imgf000004_0003

(IV) US patent No. 4,507,239 provides a process which involves the preparation of 2a-methyl-2 -azidomethylpenam derivatives of formula (VII) by treatment of compound of formula (IV) with sodium azide in aqueous aprotic solvents.

Figure imgf000005_0001

In yet another method disclosed in US patent No. 4,895,941, penam sulfoxide of formula,

Figure imgf000005_0002

(II)

wherein R represents a carboxy protecting group, is treated with 2-trimethylsilyl- 1,2,3-triazole in a sealed tube at elevated temperatures to give a mixture which upon column chromatography purification yields 2a-methyl-2 -triazolylmethyl penam derivative of formula (V).

US patent 4,518,533 provides a process for the preparation of intermediate of formula (III)

Figure imgf000005_0003

(HI) wherein the ester of penicillanic acid- 1 -oxide [compound of formula (II)] is reacted with 2-mercaptobenzothiazole in aliphatic hydrocarbon or aromatic hydrocarbon followed by isolation using column chromatographic method.

US patent 7,273,935 provides a process for the preparation of compound of formula (VIII) by reacting compound of formula (III) with cyclising agents like HCl or HBr and sodium nitrite.

Figure imgf000006_0001

(VIII)

wherein R is carboxyl protecting group and L is a leaving group like CI or Br.

US patent 6,936,711 provides a process for the preparation of protected tazobactam [compound of formula (VI)] by reacting compound of formula (VIII) with 1,2,3-triazole using a base.

In addition, US patent namely US 6,660,855, US 7,692,003, and US 7,547,777 claim process for the preparation of crystalline intermediates useful in the preparation of Tazobactam.

In general, de-protection of p-nitrobenzyl/ diphenylmethyl group in penem/penicillin core like Meropenem, Imipenem, Doripenem, Ertapenem, Faropenem, tazobactam and the like utilizes 1-10% of palladium on carbon, like commercially available 1.0%, 2.5%, 5.0%, 7.5% or 10%, which requires high pressure reactor. US patent 4,925,934 provides a de-protection method for 2a-methyl-2 – triazolylmethylpenam derivative of formula (VI) by reaction with m-cresol

Figure imgf000007_0001

(VI)

wherein R is selected from p-methoxybenzyl, diphenylmethyl (benzhydryl), 3,4,5- tirmethoxybenzyl, 2,4-dimethoxybenzyl, 3,5-dimethoxy-4-hydroxybenzyl, 2,4,6- trimethylbenzyl, ditolylmethyl, dianisylmethyl or tert-butyl. The isolated product contains higher amount of m-cresol as an impurity.

US patent 7,674,898 provides a process for the isolation of tazobactam by heating the aqueous solution containing Tazobactam before adjusting the pH. Before adjusting the pH of the aqueous solution containing tazobactam, the said solution was treated with ion-exchange resin column to purify the product. The use of ion-exchange resin and eluting the product is cumbersome on commercial scale.

Considering the importance of Tazobactam in healthcare treatment, the present inventors diligently worked to identify a robust and high yield process for the preparation of Tazobactam having cresol content below 5 ppm. A further purpose of the invention is to provide a manufacturing method that yields Tazobactam and its related intermediates with high purity and productivity.

Scheme:

Figure imgf000009_0001

Preparation of Tazobactam (I)

Into m-cresol was added benzhydryl 3-methyl-7-oxo-3-(lH-l,2,3-triazol-l- ylmethyl)-4-thia-l-azabicyclo[3.2.0]heptane-2-carboxylate 4,4-dioxide (VI) (5 g) and heated at 50-55°C till the completion of the reaction. The reaction mass was diluted with methyl isobutyl ketone. The reaction mass was extracted with sodium bicarbonate solution. The aqueous extract was acidified with hydrochloric acid to pH 3.0-4.0 and washed with methyl isobutyl ketone. Activated carbon was added, stirred and filtered. The filtrate was cooled to 0-5°C, and isopropyl alcohol (20 mL) was added followed by adjusting the pH to 1.0-2.0 using hydrochloric acid. The crystallized product was filtered, washed with water and dried.

Yield: 2.7 g

Purity: 99.9%

m-cresol content: 0.7 ppm

Example 5

Preparation of Tazobactam (I)

Into m-cresol was added benzhydryl 3-methyl-7-oxo-3-(lH-l,2,3-triazol-l- ylmethyl)-4-thia-l-azabicyclo[3.2.0]heptane-2-carboxylate 4,4-dioxide (VI) (5 g) and heated at 70-75 °C till the completion of the reaction. The reaction mass was diluted with dichloromethane. The reaction mass was extracted with potassium carbonate solution. The aqueous extract was acidified with hydrochloric acid to pH 3.0-4.0 and washed with dichloromethane. Activated carbon was added, stirred and filtered. To the filtrate, methanol (20 mL) was added followed by adjusting the pH to 1.0-2.0 using hydrochloric acid at 22-27° C. The crystallized product was filtered, washed with water and dried.

Yield : 2.6g

Purity: 99.9%

m-cresol content : 0.24 ppm

Example 6

Preparation of Tazobactam (I)

Into m-cresol was added benzhydryl 3-methyl-7-oxo-3-(lH-l,2,3-triazol-l- ylmethyl)-4-thia-l-azabicyclo[3.2.0]heptane-2-carboxylate 4,4-dioxide (VI) (5 g) and heated at 60-65 °C till the completion of the reaction. The reaction mass was diluted with dichloromethane. The reaction mass was extracted with potassium carbonate solution. The aqueous extract was acidified with hydrochloric acid to pH 3.0-4.0 and washed with dichloromethane. Activated carbon was added, stirred and filtered. To the filtrate, ethanol (20 mL) was added followed by adjusting the pH to 1.0-2.0 using hydrochloric acid at 22-27° C. The crystallized product was filtered, washed with water and dried.

Yield : 2.6g

Purity: 99.9%

m- ere sol content : 0.31

Reference example- 1

The process disclosed (example-1) in US 4,925,934 was repeated to get Tazobactam

Figure imgf000021_0001

• The above table clearly indicates that the use of water-miscible solvents helps to reduce the m-cresol content to less than 1 ppm.

• The present process obviates the use of ion-exchange resin for the purification (Refer example-1 of US 7,674,898) and provides a robust process for the industrial production of Tazobactam having less than 5ppm, preferably less than lppm. The m-cresol content in tazobactam acid is determined using HPLC with the following parameters

Colum Zorbax SB C8 (150 x 4.6mm, 3.5μ).

Mobile phase Phosphate buffer: Acetonitile

Detector UV at 200 nm

Column temperature 30°C

Flow rate 0.8 mL/min

Run time 15 min.


PATENT

CN 102020663

http://www.google.com/patents/CN102020663B?cl=en

Example 8:

 (I) in a three-necked flask were added CH2Cl2 300mL IOOOmL and 1. 5mol. [1H2SO4 lOOmL, stirring was added 81. 3g (0. 508mol) of bromine was cooled to 0 ° C after, Ilg sixteen burning trimethylammonium ammonium bromide and 35g (0. 508mol) sodium nitrite to the reaction mixture, with continuous stirring, was added portionwise 6-APA 55g (0. 254mol) and dissolved, and stirred at 0~5 ° C Ih, a solution of lmol . L-1 NaHSO3 to K1- starch paper test solution does not change color. And then allowed to stand separated, the aqueous layer was combined organic layer was extracted twice IOOmL CH2Cl2, washed successively with water, 7% aqueous NaHCO3, saturated sodium chloride aqueous solution, to give 6,6-dibromo-containing penicillanic acid in CH2Cl2 solution was used directly in the next reaction.

 (2) in IOOOmL three flask, 6,6_-dibromo penicillanic acid in CH2Cl2 solution (about 400ml), cooled to 5 ° C after the addition of benzhydrol 47g (254mmol), DCC (N, N- dicyclohexyl carbodiimide) 52. 3g (254mmol), 1. 8g of concentrated sulfuric acid was added and dissolved with stirring, at 5~10 ° C under stirring for 30min the reaction product was filtered off D⑶ DCC dehydrated to form the (N, N- two cyclohexylurea), liquor spotting, TLC [developing solvent V (cyclohexane): V (ethyl acetate) = 6: 4] to display all the raw materials after completion of the reaction on a rotary evaporator at 30~40 ° C steam dichloromethane, to give the 6,6-dibromo-penicillanic acid diphenylmethyl ester concentrate was used directly in the next reaction.

[0120] (3) obtained in the above Step 6,6-dibromo-penicillanic acid diphenylmethyl ester concentrate was added 500mL three flask, cooled with stirring to (TC, was added 0. 5g cobalt acetate Co (AC) 2 at 0~5 ° C dropping 50mL 30% H202, finished in 30min drip, drip completed at 0~5 ° C thermal reaction, TLC [developing solvent V (cyclohexane): V (ethyl acetate) = 6: 4] track to complete the reaction (about 4h). Still stratification, the organic layer was successively washed with water three times, after 7% NaHC03 was washed twice, the solvent was distilled off under reduced pressure to give 6,6-dibromo-penicillanic alkylene acid diphenylmethyl ester sulfoxide The crude product without purification, was used directly in the next reaction.

 (4) the 6,6-dibromo-penicillanic acid diphenylmethyl ester sulfoxide The crude product was dissolved in 4001 ^ tetrahydrofuran, at 101: add 150mL 10% NH4Cl solution, zinc powder was added in four portions 82. 5g (127mol), at intervals IOmin, about 50min addition was completed, plus complete response at 0~10 ° C 30min. Plus zinc filtered through Celite, standing stratified rotating concentrated organic layer recovered tetrahydrofuran. Ethyl acetate was added to dissolve the concentrated solution, washed with water, saturated aqueous sodium chloride solution, dried over anhydrous magnesium sulfate, concentrated under reduced pressure (45 ° C or less) to just precipitate a solid, 0~5 ° C curing crystallization 3h, suction and the filter cake was dried in vacuo to give white crystals of 6,6-dihydro-penicillanic acid sulfoxide, diphenylmethyl ester 70g, 72% yield [6-APA to calculate, yield = weight of dry product / (6-APA was mass X 383)], mp 145 ~147 ° C (literature value of 145 ~148 ° C).

 (5) containing 6,6-dihydro-penicillanic acid sulfoxide, diphenylmethyl ester (70g, 0. 182mol), 2- trimethylsilyl-1,2,3-triazole (25. 7g, the 0. 182mol) and toluene (500mL) autoclave purged with nitrogen, then heated to 110~120 ° C, the reaction 4.5h. After cooling, toluene was evaporated, extracted with ethyl acetate (700mL), water (250mL) washed with saturated sodium chloride solution (250mL), dried over anhydrous magnesium sulfate, the solvent was evaporated, and recrystallized from ethanol to give 2a- A yl 23- (1,2,3-triazol-1-yl) methyl penicillanate -3 a- carboxylic acid, diphenylmethyl ester (white solid) 43.48g, 55% yield [yield = dry product Weight / (() • 182X434. 4)], mp 140 ~142 ° C (literature values ​​141 ~143 ° C).

 (6) The 2a- methyl 2 P – (1,2,3`_ triazol _1_ yl) methyl penicillanate _3 a – carboxylic acid diphenylmethyl ester 43. 48g (0.1OOmoI ) was dissolved in 35mL of acetone, was added 70mL of water and 105mL of glacial acetic acid, cooled to 0~5 ° C, was added with stirring a mixture of KMnO4 (23. 7g KMnO4,16. 5g of concentrated phosphoric acid, and 520ml water), with 5mol. L- phosphate, pH was adjusted to 1 6.5, the reaction was stirred at room temperature for 3h. 30% hydrogen peroxide was added dropwise to the reaction solution colorless, filtered, and the resulting crude product was recrystallized from methanol to give 40. 6g as a white solid (2 a- methyl 2 ¢ – (1, 2,3- triazol-1-yl) methylpenicillanate _3 a – carboxylic acid diphenylmethyl ester-dioxide), 87% yield [yield = weight of dry product / (0 100X466.7).]. mp 205~207 ° C (206 ~208 literature values ​​..).

 (7) in 500ml reaction flask was added 2 a- methyl 2 ¢ – (1, 2,3- triazol-1-yl) methyl penicillanate _3 a- two carboxylic acid diphenylmethyl ester oxide 40. 6g (0. 087mol) and 200mL (2mol) between A sprinkle, stirred until solid was completely dissolved, 80 ° C the reaction was kept 4h, cooled to room temperature, was added 600mL of methyl isobutyl ketone, with IOOmL 7% carbonate solution of sodium hydroxide wash, the aqueous layer was separated, the organic layer was washed twice with 150ml, the combined aqueous layer was cooled to 0~5 ° C, with 6mol. L-1 hydrochloric acid to adjust the pH to I~1. 8, white crystals precipitated, pumping filter, 80 ° C drying, dry goods tazobactam 15. 2g, 58% yield [yield = dry goods weight / (0. 087X300. 3)] o mp 136 ~137 ° C (literature value of 136 ~ 138 ° C).

Figure CN102020663BD00061

 PATENT

http://www.google.com/patents/EP1686131A2?cl=en

  • methods of producing β-substituted methyl penam derivatives. For instance, US 4,529,592 discloses a process which involves the treatment of 2α-methyl-2β-azidomethyl penam derivatives of formula (c):

    Figure imgb0003

    wherein R is a carboxy-protecting group, with acetylene, an acetylene derivative or a vinyl derivative under high pressure in a sealed reactor and at elevated temperatures, followed by deprotection with a suitable reagent to get the β-lactamase inhibitor of formula (a).

  • The 2α-methyl-2β-azidomethyl penam derivative of formula (c) is in turn prepared from the 2α-methyl-2β-halomethyl penam derivatives of formula (d)

    Figure imgb0004

    wherein R is a carboxy-protecting group and X is chloro or bromo, by treating with sodium azide in aqueous polar aprotic solvents, followed by oxidation.

  • US 4,891,369 and US 4,933,444 disclose a different approach, which involves the preparation of 2α-methyl-2β-triazolylmethylpenam derivatives of formula (e) wherein R is a carboxy protecting group and n is 0, by the treatment of a β-halomethyl penam derivative of formula (d), wherein X is chlorine or bromine and R is a carboxy-protecting group, with 1H-1,2,3-triazole.

    Figure imgb0005

    The product obtained can be oxidized and deprotected to get the 2β-substituted methyl penam compound (a).

  • US 4,912,213 discloses a reduction method employing lead salts in catalytic amounts to prepare a 2α-methyl-2β-triazolylmethyl penam derivative of formula (e) (n=0-2) from 6-halo or 6,6-dihalo-2α-methyl-2β-triazolylmethyl penam derivatives of formula (f)

    Figure imgb0006

    where X may be Cl, Br, I; Y may be Cl, Br, I or a hydrogen atom; and R is a carboxy-protecting group.

  • In yet another method disclosed by US 4,895,941, penam sulfoxide of formula (g), wherein R represents a carboxy-protecting group, is treated with 2-trimethylsilyl-1,2,3-triazole in a sealed tube at elevated temperatures to give a mixture which requires purification by column chromatography to isolate the 2α-methyl-2β-triazolylmethyl penam derivative of formula (e) (n=0).

    Figure imgb0007
  • As an alternative to the hydrogenation, US 4,925,934 discloses a deblocking method for a 2α-methyl-2β-triazolylmethyl penam derivative of formula (h) by reaction with cresol

    Figure imgb0008

    where R is selected from p-methoxybenzyl, 3,4,5-trimethoxybenzyl, 2,4-dimethoxybenzyl, 3,5-dimethoxy-4-hydroxybenzyl, 2,4,6-trimethylbenzyl, diphenylmethyl, ditolylmethyl, dianisylmethyl or tert-butyl.

  • [0009]

    Published application US 2003/232983 discloses a complete different route of synthesis for 2α-methyl-2β-triazolylmethyl-penam derivatives starting from cepham derivates of formula (i) by substitution and rearrangement

    Figure imgb0009

    where R represents a carboxy-protecting group and L a leaving group.

  • In most of the methods involved, 2α-methyl-2β-halomethyl penam of formula (d) is used as the key intermediate. This is true with both the azide/acetylene combo and the triazole route discussed above. However, the 2α-methyl-2β-halomethyl penam of formula (d) itself is an unstable intermediate and therefore manufacturing and storage of this intermediate in large quantities is always cumbersome. This intermediate has been found to degrade on storage even at low temperatures in isolated form as well as in the solution from which it is isolated. Thus, all the operations related to preparation of the intermediate have to be done rapidly, and the isolated intermediate has to be converted to the final product immediately. As a result of these limitations, in-plant scale up always yields by-products which ultimately require purification demands.

Example 1: Preparation of Tazobactam Sodium by route A. (Fig. 1)Step 1.

      Production of 6α-Bromopenicillanic acid (BPA) (compound II)

    • 2.5 L of 1.24 molar sulphuric acid (3.125 mol) was stirred at 4°C in a 6 L flask. 218.4 g (1.0 mol) of 6-APA (99%) (compound I) following 601 g (5.05 mol) of potassium bromide and 2000 mL of ethanol were added, maintaining the temperature between 4 to 8°C. Inorganic salts were removed by filtration. The resulting cake was washed by 2 x 1.25 L of cooled dichloromethane. The aqueous phase was extracted twice using the previous washing liquor and 3 x 500 mL of cooled dichloromethane. The organic phases were combined (approx. 4.0 L) and washed with 2 x 200 mL of 30% brine at 4°C. The greenish-brown solution was concentrated to 700 mL in vacuum. The precipitate was removed by filtration and the solution was kept below 0°C and used without further purification in the next reaction step.
      Yield: 90% (by titration)
      TLC (thin layer chromatography; detection by UV and phosphomolybdic acid, eluent: acetone – methanol 2:1 v/v): Rf 0.65 (BPA), (eluent: acetone – methanol 4:1 v/v) Rf 0.35 (BPA)

Step 2

      . Production of 6α-Bromopenicillanic acid-S-oxide (BPO) (compound III)

    • 1.8 mol of BPA in 1400 mL of dichloromethane was placed in a 4 L flask. The temperature of the solution was maintained between 0 to 2°C. 2.0 mol peracetic acid in acetic acid solution (342 mL, 40 wt.-% peracetic acid) was added within 100 to 120 minutes, maintaining the temperature of the solution between 0 to 8°C. The color of the solution changed to yellowish-brown. The solution was stirred further 1 hour at 0 to 8°C. The product crystallizes. The slurry was cooled to -10 to -15°C and stirred further 30 minutes then filtered. The cake was washed with 2 x 400 mL of dichloromethane at -10°C. The product was dried at 20 – 25°C in vacuum. The crude product was kept below 0°C and used without further purification immediately (storage time 1 to 2 days) in the next reaction step.
      Yield: 314 – 331g (58,9 – 62.1 %)    Mp: 130 °C (decomp.)
      Cumulative yield of 1st and 2nd steps: 51- 52%
      TLC (detection by UV and phosphornolybdic acid, eluent: acetone – methanol 2:1 v/v)
      Rf 0.65 (BPA), Rf 0.45 (BPO)
      The yield can be improved using higher concentrated peracetic acid.

Step 3

      . Production of 6α-Bromopenicillanic acid-S-oxide p-nitrobenzyl ester (BPE) (compound IV)

    • In a 4 L flask 272.44 g (0.92 mol) of BPO was dissolved in 120 mL DMF at 25°C. 100.8 g (1,2 mol) of sodium hydrogencarbonate and 229.0 g (1.06 mol) of p-nitrobenzylbromide (PNM) were added portionwise. The slurry was cooled and stirred at 0 to 5°C for one hour. The product was filtered and washed with 2 x 800 mL of cold water. The wet product was placed in a 2 L flask and 1200 mL of methanol was added. The slurry was refluxed for one hour, cooled to -10°C and filtered. The cake was washed with 2 x 800 mL of methanol at -10°C. The product was dried at 25 – 30°C in vacuum and stored at 0°C without further purification in the next reaction step.
      Yield: 334.8 g (84.4%)    Mp: 130 °C (decomp.)
      Cumulative yield of 1st, 2nd and 3rd steps: 46%
      TLC (detection by UV, eluent: acetone – methanol 2:1 v/v) Rf 0.75 (BPE), Rf 0.65 (BPO);
      (eluent: ethyl acetate – hexane 2:1 v/v) Rf 0.50 (BPE), Rf 0.00 (BPO)

Step 4.

      Production of 2-(2-Benzothiazolyldithio)-3-bromo-α-(1-methylethylidene)-4-oxo-1-azetidincacetic acid p-nitrobenzyl ester (BBE) (compound V)

    • In a 4 L flask 140.84 g (0.826 mol) of 95% 2-mercaptobenzothiazole (MBT) and 345.0 g BPE (0.8 mol) were dissolved in 1360 mL toluene when the solution was heated to 86 – 90°C and an azeotropic mixture of toluene-water was distilled at 450 to 500 mbar. After 3 to four hours, 14 to 16 mL of water was removed using a Dean-Stark apparatus maintaining the temperature between 86 to 90°C. If unreacted BPE could be detected by TLC, a small amount of 2 to 8 g of MBT was added. The solution was refluxed until no starting material could be detected by TLC.
    • The solution was evaporated in vacuum between 60 to 70°C. The residual oil was dissolved in 1200 mL of ethyl acetate. After cooling the product crystallizes. The slurry was concentrated in vacuum below 50 °C to 800 mL and 1200 mL isopropyl ether was added to give a well-filterable crystalline slurry that was cooled below 20°C and stirred for additional 24 hours. Subsequently, the product was filtered and washed with 2 x 500 mL cooled isopropyl ether. The product was dried in vacuum between 25 – 30°C.
      Yield: 412.8 g (88.9%)    Mp.: 116-119°C
      Cumulative yield of 1st, 2nd, 3rd and 4th steps: 41%
      TLC (detection by UV, eluent: isopropyl ether – ethyl acetate 99:1 v/v) Rr 0.65 (BBE)

Step 5

      . Production of 6α-Bromo-2β-bromomethyl-2α-methylpenam-3α carboxylic acid p-nitrobenzyl ester (DBPE) (compound VI)

    • In a 4 L flask 290.24 g (0.5 L) of BBE was dissolved in 1500 mL dichloromethane. The solution was cooled to -2°C. 540 mL of 30% aqueous solution of hydrogen bromide (2.52 mol) was added, keeping the temperature below 0°C. A solution of 103.5 g (1.5 mol) sodium nitrite in 300 mL was added keeping the temperature between 0 to 3 °C. Meanwhile the colour of the organic phase turned to brown. The reaction mixture was stirred about 90 min at 0 to 5 °C until the starting material could not be detected by TLC. 80 g of sodium carbonate (0.75 mol) was added, adjusting the pH to between 6 and 7. The reaction mixture was filtered using perlite as a filter aid. The precipitate was washed with 3 x 100 mL dichloromethane. The combined organic layer was separated and concentrated to 700 mL. The solution was cooled to 20 °C and two litres of isopropyl ether were added slowly. The crystalline suspension was stirred 16 hours at 20 °C and two hours at 0 °C. It was filtered and the product was washed with 2 x 300 mL of cooled isopropyl ether. The product was dried at 20 to 25 °C in vacuum.
      Yield: 235.84 g (95.5%)    Mp.: 80 °C (decomp.)
      Purity: min. 95 %
      Cumulative yield of 1st – 5th steps: 39%
      The product is sensitive to light and decomposes on silica gel to give cepham.
      TLC (detection by UV, eluent: isopropyl ether – ethyl acetate 99:1 v/v) Rf 0.72 (DBPE),
      Rf 0.65 (BBE), Rf 0.57 (cepham)

Step 6

      . Production of 6α-Bromo-2β-azidomethyl-2α-methylpenam-3α-carboxylic acid p-nitrobenzyl ester (BTPE) (compound VII)

    • In a 2 L flask 292.3 g (342 mL, 2.664 mol) trimethylsilylchloride was dissolved in 1300 mL of toluene. 210.1 g (3.20 mol) sodium azide was added and the suspension was stirred and refluxed. The reaction was traced by GC. After 10 to 16 hours less than 0.1% of the starting material could be detected. The suspension was cooled to -5 to 0°C and was filtered (or decanted). The solution (1580 mL) contains 2.40 mol of trimethylsilylazide, which is volatile (Bp: 95°C) and a toxic compound.
    • In a 2 L flask 52.63 g (23.7 mL, 0.2 mol) tin(IV) chloride was added to a toluene solution of 2.4 mol of trimethylsilylazide between 20 – 25°C. The solution was stirred 24 hours at 20 – 25 °C while some white precipitate appeared. 197.7 g (0.4 mol) DBPE was added. The suspension was stirred 40 to 70 hours while brown gum appeared. The formation of azide was traced by TLC (eluent isopropyl ether – ethyl acetate 99:1 v/v) Rf 0.72 (DBPE), Rf 0.61 (BAPE), Rf 0.58 (cephambromide) Rf 0.40 (cephamazide).
    • Conversion of the starting material to product was less than 50% after 40 hours. Additionally, 0.2 mol of tin (IV) chloride was added, which accelerated the formation of BAPE.
    • After no starting material could be detected by TLC, the reaction mixture was quenched with 1200 mL of saturated sodium carbonate solution at 5-10°C. The insoluble material was dissolved by 400 mL ethyl acetate and added to the sodium carbonate solution. The biphasic reaction mixture was stirred 15 minutes, The pH of the lower aqueous phase was between 8 and 9. Perlite (50 g) as a filter aid was added and the suspension was filtered. The cake was washed with 2 x 200 mL of ethyl acetate.
    • The combined filtrates were poured into a 5 L separating funnel and the lower aqueous phase was removed and extracted with 2 x 200 mL ethyl acetate. The combined organic phases were washed by 200 mL saturated sodium bicarbonate solution and 200 mL brine. The solvent was removed in vacuum and the residue was suspended in 1000 mL methanol at 0 – 5 °C. The crystalline suspension was stirred 2 to 3 hours at 0 – 5 °C and filtered. The product was washed with 200 mL diisopropyl ether and dried in vacuum at 20 – 25 °C.
      Yield: 153.8 g (84.3%)
      Purity: 68 ― 70% (by HPLC: mobile phase 0.05 M KH2PO4 – acetonitrile 1:1, pH 6,
      Rf 14.33 min)
      Cumulative yield of 1st – 6th steps: 33%

Step 7

      . Production of 6α-Bromo-2β-[(1,2,3-triazol-1-yl)methyl]-2α-methylpenam-3α-carboxylic acid p-nitrobenzyl ester (BTPE) (compound VIII)

    • In a 1 L autoclave 7.6 g (50 mmol) BAPE was dissolved in 640 mL 2-butanone. The solution was cooled down to 0 – 5 °C. The autoclave was pressured three times with nitrogen gas up to six bar. The autoclave was filled with acetylene gas up to 1.5 bar pressure and approx. 36 g acetylene gas was dissolved. The autoclave was heated gradually from 0 °C up to 84 – 94 °C, keeping the pressure between 5 – 6 bar. The reaction mixture was stirred in the autoclave 14 – 20 hours at 84 to 94 °C and pressure of 5 to 6 bar. No starting material was detected by TLC (eluent hexane – ethyl acetate 1:2 v/v) Rf> 0.9 (BAPE), Rf 0.51 (BTPE), Rf 0.32 (cephamtriazole).
    • The autoclave was cooled down to -20 to -25 °C and 7.6 g BAPE in 50 mL 2-butanone solution was added. The autoclave was heated again to 84 – 94 °C and the reaction mixture was stirred 14 to 20 hours at 84 – 94 °C. The autoclave was cooled and the procedure was repeated with 7.6 g BAPE. The autoclave was cooled down to 20 – 25 °C and opened. The reaction mixture was poured into a 1 L flask and was concentrated in vacuum up to 140 mL. The solution was cooled to 0 – 5°C. The crystalline suspension was stirred for 1 hour and was filtered. The product was washed with 40 mL cool 2-butanone. The product was dried in vacuum at 25 – 30 °C.
      Yield: 13.51 g (56.0%)    Mp.: 180-182°C (decomp.)
      Purity: 98.6% (by HPLC: mobile phase 0.05 M KH2PO4 – acetonitrile 1:1, pH 6,
      Rf 8.40 min)
      Cumulative yield of 1st– 7th steps: 18%

Step 8

      . Production of p-Nitrobenzyl 6α-bromo-2α-methyl-2β-(1,2,3-triazol-1-yl)methylpenam-3α -carboxylate-1,1-dioxide (compound. IX)

    • To a solution of 4.82 g (10.00 mmol) of BTPE in a mixture of 210 ml of acetic acid and 27 ml of water, 3.79 g (23.6 mmol) of KMnO4 was added in 30 minutes at room temperature. The progress of the reaction was monitored by TLC. When the reaction was complete, the excess of KMnO4 was destroyed by 30 % H2O2 solution. The reaction mixture was poured into 930 mL of cold water, the precipitated product was filtered and washed with cold water and dried over P2O5, giving compound IX.
      Yield: 4,12 g (80 %)
      Purity: more than 95 % (HPLC)    Mp.: 122-124°C
      TLC (detection by UV, eluent: ethyl acetate – hexane 2:1 v/v) Rf 0.51 (VIII), Rf 0.23 (IX)

Step 9

      . Production of Tetrabutylammonium 2α-methyl-2β-(1,2,3-triazol-1-yl)methylpenam-3α -carboxylate-1,1-dioxide (compound Xa)

    • A stainless steel stirred autoclave with a total volume of 1 L was charged with 5.1 g (10 mmol) of compound IX, 2.5 g (30 mmol) of NaHCO3, 1.0 g of 10 % Pd on charcoal, 100 mL of water and 100 mL of ethyl acetate. The autoclave was sealed and flushed with argon, then pressured with hydrogen up to 14 bars. The hydrogenation was carried out at room temperature for 5 h. Completion of the reaction was checked by TLC. The mixture was filtered and the filter washed with water. The aqueous phase was separated, washed with ethyl acetate (2 × 10 mL) and Bu4NNaSO4 solution (prepared from 340 mg (1 mmol) of Bu4NHSO4 and 84 mg (1 mmol) of NaHCO3 in 5 mL of water) added. The aqueous solution was extracted with dichloromethane (5 x 10 ml). The combined dichloromethane phases were dried over Na2SO4 and concentrated under reduced pressure to dryness keeping the temperature of the water bath below 20 °C.
      Yield: 0.39 g (75 %)
      Purity: 95.5 % (HPLC)
      HPLC mobile phase: 0.05 M KH2PO4 buffer, pH 2.3
      Eluent A: 95 % of 0.05 M KH2PO4 buffer (pH 2.3) plus 5 % acetonitrile
      Eluent B: 40 % of 0.05 M KH2PO4 buffer (pH 2.3) plus 60 % acetonitrile
      Retention time: 11.53 min
      Column: RP-18 endcapped (5µm, 250 mm)
      TLC (detection by UV and 1 % AgNO3 in ethanolic solution, eluent: ethyl acetate – hexane 2:1 v/v) Rf 0.23 (IX); (eluent: acetone -methanol 2:1 v/v) Rf 0.48 (Xa)

Step 10.

    Production of Sodium 2α-methyl-2β-(1,2,3-triazol-1-yl)methylpenam-3α-carboxylate-1,1-dioxide (Tazobactam sodium)

  • The residue containing compound Xa (0.40 g) was eluted with water on a column of Amberlite-Na+ cation-exchange resin. The appropriate fractions were concentrated under reduced pressure and finally lyophilized, yielding Tazobactam sodium.
    Yield: 0.21 g (85 %)
    Purity: 99.5 % (HPLC)
    HPLC mobile phase: 0.05 M KH2PO4 buffer, pH 2.3
    Eluent A: 95 % of 0.05 M KH2PO4 buffer (pH 2.3) plus 5 % acetonitrile
    Eluent B: 40 % of 0.05 M KH2PO4 buffer (pH 2.3) plus 60 % acetonitrile
    Retention time: 11.53 min
    Column: RP-18 cndcapped (5µm, 250 mm)

PATENT

http://www.google.com/patents/US8476425

Tazobactam arginine can be a salt consisting of the conjugate base of (2S,3S,5R)-3-((1H-1,2,3-triazol-1-yl)methyl)-3-methyl-7-oxo-4-thia-1-azabicyclo[3.2.0]heptane-2-carboxylic acid 4,4-dioxide (tazobactam) and the conjugate acid of (S)-2-amino-5-guanidinopentanoic acid (L-arginine) in a 1:1 ratio, as represented by the structure below.

Figure US08476425-20130702-C00001

References

  1. Yang Y, Rasmussen BA, Shlaes DM (1999). “Class A beta-lactamases—enzyme-inhibitor interactions and resistance”. Pharmacol Ther. 83: 141–151. doi:10.1016/S0163-7258(99)00027-3.
CN1037514A Mar 1, 1989 Nov 29, 1989 大鹏药品工业株式会社 Process for preparing 2 alpha-methyl-2 beta-(1,2,3-triazole-1-yl) methylpenam-3 alpha-carboxylic acid derivatives
US7674898 * Jul 23, 2001 Mar 9, 2010 Otsuka Chemical Co., Ltd. Anhydrous crystal of β-lactam compound and method for preparation thereof
Reference
1 * LI YANG ET AL.: ‘Synthesis of Tazobactam, [beta- Lactamase Inhibitor‘ TRANSACTIONS OF TIANJIN UNIVERSITY vol. 8, no. 1, March 2002, pages 33 – 36


Tazobactam
Tazobactam.svg
Tazobactam ball-and-stick.png
Systematic (IUPAC) name
(2S,3S,5R)-3-Methyl-7-oxo-3-(1H-1,2,3-triazol-1-ylmethyl)-4-thia-1-azabicyclo[3.2.0]heptane-2-carboxylic acid 4,4-dioxide
Clinical data
AHFS/Drugs.com International Drug Names
Pregnancy
category
  • B
Legal status
  • (Prescription only)
Routes of
administration
Intravenous
Identifiers
CAS Number 89786-04-9 Yes
ATC code J01CG02
PubChem CID: 123630
DrugBank DB01606 Yes
ChemSpider 110216 Yes
UNII SE10G96M8W Yes
KEGG D00660 Yes
ChEBI CHEBI:9421 Yes
ChEMBL CHEMBL404 Yes
Chemical data
Formula C10H12N4O5S
Molecular mass 300.289 g/mol
Patent Submitted Granted
2-OXO-1-AZETIDINE SULFONIC ACID DERIVATIVES AS POTENT BETA-LACTAMASE INHIBITORS [EP0979229] 2000-02-16 2002-10-23
DHA-pharmaceutical agent conjugates of taxanes [US7199151] 2004-09-16 2007-04-03
Antimicrobial composition comprising a vinyyl pyrrolidinon derivative and a carbapenem antibiotic or a beta-lactamase inhibitor [EP0911030] 1999-04-28 2005-04-13
7-alkylidene-3-substituted-3-cephem-4-carboxylates as beta-lactamase inhibitors [US7488724] 2006-04-06 2009-02-10
Sustained release of antiinfectives [US7718189] 2006-04-06 2010-05-18
Conjugate of fine porous particles with polymer molecules and the utilization thereof [US2006159715] 2006-07-20
ENGINEERED BACTERIOPHAGES AS ADJUVANTS FOR ANTIMICROBIAL AGENTS AND COMPOSITIONS AND METHODS OF USE THEREOF [US2010322903] 2009-01-12 2010-12-23
Microparticles for the treatment of disease [US2010323019] 2010-08-19 2010-12-23
Packaging System [US2010326868] 2010-08-30 2010-12-30
COMBINATION ANTIBIOTIC AND ANTIBODY THERAPY FOR THE TREATMENT OF PSEUDOMONAS AERUGINOSA INFECTION [US2010272736] 2010-02-04 2010-10-28
Citing Patent Filing date Publication date Applicant Title
CN102304139A * Jul 12, 2011 Jan 4, 2012 景德镇市富祥药业有限公司 Method for preparing 2 beta-methyl penicillanate benzhydryl dioxide
CN102304139B Jul 12, 2011 Jun 4, 2014 江西富祥药业股份有限公司 Method for preparing 2 beta-methyl penicillanate benzhydryl dioxide
CN102382123A * Mar 10, 2011 Mar 21, 2012 海南美好西林生物制药有限公司 Preparation method of tazobactam sodium
CN102827189A * Sep 18, 2012 Dec 19, 2012 山东罗欣药业股份有限公司 Tazobactam sodium compound and pharmaceutical composition thereof
US8476425 Sep 27, 2012 Jul 2, 2013 Cubist Pharmaceuticals, Inc. Tazobactam arginine compositions
US8906898 May 28, 2014 Dec 9, 2014 Calixa Therapeutics, Inc. Solid forms of ceftolozane
US8968753 May 22, 2014 Mar 3, 2015 Calixa Therapeutics, Inc. Ceftolozane-tazobactam pharmaceutical compositions
US9044485 Apr 11, 2014 Jun 2, 2015 Calixa Therapeutics, Inc. Ceftolozane antibiotic compositions

SEE BACTAM SERIES…………..http://apisynthesisint.blogspot.in/p/bactam-series.html

/////////

O=S2(=O)[C@]([C@@H](N1C(=O)C[C@H]12)C(=O)O)(Cn3nncc3)C

or

CC1(C(N2C(S1(=O)=O)CC2=O)C(=O)O)CN3C=CN=N3

NXL104, Avibactam


NXL-104, Avibactam

trans-7-oxo-6-(sulphooxy)-1,6-diazabicyclo[3,2,1]octane-2-carboxamide sodium salt (e.g., NXL-104)

CAS 396731-20-7, 1192491-61-4

AVE-1330
AVE-1330A

PHASE 1 a broad-spectrum intravenous beta-lactamase inhibitor, was under development for the treatment of infections due to nosocomial drug resistant Gram-negative bacteria

SANOFI  INNOVATOR

Novexel holds exclusive worldwide development and commercialization rights from Sanofi.

NXL104; Avibactam; UNII-7352665165;

Molecular Formula: C7H11N3O6S
Molecular Weight: 265.24374 g/mol

CAS 1192500-31-4, 396731-14-9

[(2S,5R)-2-carbamoyl-7-oxo-1,6-diazabicyclo[3.2.1]octan-6-yl] hydrogen sulfate

(2S,5R)-7-oxo-6-(sulfooxy)-1,6-diazabicyclo[3.2.1]octane-2-carboxamide

trans-7-oxo-6-(sulfooxy)-1,6-diazabicyclo[3.2.1]octan-2-carboxamide

1,6-Diazabicyclo(3.2.1)octane-2-carboxamide, 7-oxo-6-(sulfooxy)-, (1R,2S,5R)-rel-

Launched-2015, Avycaz  Zavicefta, CAZ-104, CAZ-AVI
NXL-104/ceftazidime, Cephems (Cephalosporins), Fixed-Dose Combination
Avibactam is a non-β-lactam β-lactamase inhibitor that is available in combination with ceftazidime (Avycaz). This combination was approved by the FDA on February 25, 2015 for the treatment of complicated intra-abdominal infections in combination with metronidazole, and the treatment of complicated urinary tract infections, including pyelonephritis caused by antibiotic resistant-pathogens, including those caused by multi-drug resistant gram-negative bacterial pathogens. As there is limited clinical safety and efficacy data, Avycaz should be reserved for patients over 18 years old who have limited or not alternative treatment options.
Image result for AVIBACTAM

Avibactam is a non-β-lactam β-lactamase inhibitor antibiotic being developed by Actavis jointly with AstraZeneca. A new drug application for avibactam in combination with ceftazidime was approved by the FDA on February 25, 2015, for treating complicated urinary tract and complicated intra-abdominal Infections caused by antibiotic resistant-pathogens, including those caused by multi-drug resistant gram-negative bacterial pathogens.[2][3][4]

Increasing resistance to cephalosporins among Gram-(-) bacterial pathogens, especially among hospital-acquired infections, results in part from the production of beta lactamase enzymes that deactivate these antibiotics. While the co-administration of a beta lactamase inhibitor can restore antibacterial activity to the cephalorsporin, previously approved beta lactamase inhibitors such astazobactam and Clavulanic acid do not inhibit important classes of beta lactamase including Klebsiella pneumoniae carbapenemases (KPCs), metallo-beta lactamases, and AmpC. Avibactam inhibits KPCs, AmpC, and some Class D beta lactamases, but is not active aganist NDM-1.[5]

U.S. Pat. No. 7,112,592 discloses novel heterocyclic compounds and their salts, processes for making the compounds and methods of using the compounds as antibacterial agents. One such compound is sodium salt of trans-7-oxo-6-(sulphooxy)-1,6-diazabicyclo[3,2,1]octane-2-carboxamide. Application WO 02/10172 describes the production of azabicyclic compounds and salts thereof with acids and bases, and in particular, trans-7-oxo-6-sulphoxy-1,6-diazabicyclo[3.2.1]octane-2-carboxamide and its pyridinium, tetrabutylammonium and sodium salts. Application WO 03/063864 and U.S. Patent Publication No. 2005/0020572 describe the use of compounds including trans-7-oxo-6-(sulphooxy)-1,6-diazabicyclo[3,2,1]octane-2-carboxamide sodium salt, as β-lactamase inhibitors that can be administered alone or in, combination with β-lactamine antibacterial agents. These references are incorporated herein by reference, in their entirety.

str6

Figure

ChemSpider 2D Image | avibactam sodium | C7H10N3NaO6S


INGREDIENT UNII CAS Average: 287.22

C7H10N3NaO6S

Avibactam sodium 9V824P8TAI 1192491-61-4
sodium (1R,2S,5R)-2-carbamoyl-7-oxo-1,6-diazabicyclo[3.2.1]octan-6-yl sulfate
Sulfuric Acid Mono[(1R,2S,5R)-2-(aMinocarbonyl)-7-oxo-1,6-diazabicyclo[3.2.1]oct-6-yl] Ester SodiuM Salt
UNII:9V824P8TAI
UNII-C8SM6IRW7G
({[(2S,5R)-2-Carbamoyl-7-oxo-1,6-diazabicyclo[3.2.1]oct-6-yl]oxy}sulfonyl)oxydanide de sodium [French][ACD/IUPAC Name]
(1R,2S,5R)-2-carbamoyl-7-oxo-1,6-diazabicyclo[3.2.1]octane-6-yl sodium sulfate
1,6-Diazabicyclo[3.2.1]octane-2-carboxamide, 7-oxo-6-(sulfooxy)-, sodium salt, (2S,5R)- (1:1) [ACD/Index Name]


Avibactam, sodium (2S,5R)-2-carbamoyl-7-oxo-1,6-diazabicyclo[3.2.1]octan-6-yl sulfonate, containing a diazabicyclo[3.2.1]octane (DBO) heterocyclic core structure, is a novel diazabicyclooctane non-β-lactama β-lactamase inhibitor. It has a unique mechanism of inhibition among β-lactamase inhibitors, which is able to bind reversibly and covalently to β-lactamase.

As a new drug featured with bacterial resistance, avibactam has been widely used in clinic and its combination with Ceftazidime (Zavicefta) has recently been approved by the EMA and FDA for treatment of complicated intra-abdominal infectious (CIAI), complicated urinary tract infectious (CUTI), hospital acquired pneumonia (HAP), etc.

Moreover, compared with the three known β-lactamase inhibitors named clavulanic acid, sulbactam, and tazobactam, the efficiency of avibactam is stronger and its spectrum is also broader: avibactam is active against class A including Class A Klebsiella pneumoniae carbapenemase (KPCs) and ESBLs, class C, and some class D β-lactamases

Image result for AVIBACTAM

Synthesis

SYN1

Figure

Substantial effort has been devoted to the preparation of avibactam . Initially, the Aventis infection division in Romainville (France) disclosed route A for its synthesis in the early stage of drug discovery. In this process, double-chiral piperidine derivatives were used as starting material to provide avibactam via inversion of configuration, deprotection, urea-cyclization, deprotection, and sulfonation with about 9.0% total yields. Miller et al. also prepared avibactam based on route A. This route suffers from a long synthetic procedure, low yield, and heavy laborious workups. Besides, the raw materials (double-chiral piperidine derivatives) are expensive and a number of environmentally undesirable reagents and solvents are required in this route.

  • DubreuilL. J.MahieuxS.NeutC.MiossecC.PaceJ. Int. J. Antimicrob. Agents 201239 (6500– 504 DOI: 10.1016/j.ijantimicag.2012.02.013

  • (a) MangionI. K.RuckR. T.RiveraN.HuffmanM. A.ShevlinM. Org. Lett. 201113 (205480– 5483 DOI: 10.1021/ol202195n

    (b) KrishnamurthyS.VenkataprasadJ.VagvalaT. C.MoriguchiT.TsugeA. RSC Advances, 5 (64), 52154– 52160.

  • (a) Aventis Pharma SA: WO2002010172, 2002.

    (a) AszodiJ.LampilasM.FromentinC.RowlandsD. A. Aventis Pharma SAFR, 2835186 (2003) .

    (c) Novexel: WO2008142285, 2008.

    BaldwinJ. E.AdlingtonR. M.GodfreyC. R.GollinsD. W.SmithM. L.RusselA. T. Synlett 19931993 (0151– 53 DOI: 10.1055/s-1993-22345

The Wockhardt developed route B to obtain avibactam from l-glutamate acid or l-pyroglutamic acid. In this route, the skeleton of the target molecular diazabicyclo[3.2.1]octane heterocyclic core structure (DBO) was constructed through the steps of ring-opening, ring-closing, deoxygenization, and then by deprotection, sulfonation, and other steps to obtain avibactam with a total yield of about 11.0%. This method, producing small scale avibactam in a single batch, has some drawbacks limiting the large-scale synthesis: (a) a long synthetic procedure, (b) complicated purification process, and (c) the employment of excessive environmentally unfriendly reagents such as diphosgene.

(a) PatilV. J.TadiparthiR.BirajdarS.BhagwatS. P. US8969334, 2015.

(b) HeckerE. A.BaldwinA. B-lactamase inhibitor picoline salt: P, US9120796, 2015.

(c) AszodiJ.FromentinC.LampilasM.RowlandsD. A. P. US7612087, 2009.

(d) GuY. G.HeY.YinN.AlexanderD. P. WO2013149136, 2013.

(e) HwangY. S.GuJ. Q.JainA.GaradS.JacobP. S. P. US20140275001, 2014.

(f) HeckerE. A.BaldwinA. P. US20159120796, 2015.

Recently, AstraZeneca and Forest Laboratories have optimized the process: from commercially available Boc-benzylglutamate in only 5 isolated steps with an overall yield of 35.0% (without including the construction of DBO). Another route is based on the olefin metathesis reaction to construct the DBO skeleton (Route C)

XiongH.ChenB.Durand-RévilleT. F.JoubranC.AlelyunasY. W.WuD.HuynhH. ACS Med. Chem. Lett. 20145 (101143– 1147DOI: 10.1021/ml500284k

BallM.BoydA.EnsorG. J.EvansM.GoldenM.LinkeS. R.MilneD.MurphyR.TelfordA.KalyanY.LawtonG. R.Rachas.ZhouS. H. Org. Process Res. Dev. 201620 (101799– 1805 DOI: 10.1021/acs.oprd.6b00268

SYN2

Org. Process Res. Dev., Article ASAP
DOI: 10.1021/acs.oprd.7b00290
Publication Date (Web): December 7, 2017
STR1

A New Synthetic Route to Avibactam: Lipase Catalytic Resolution and the Simultaneous Debenzylation/Sulfation

 Research & Development Center, Zhejiang Medicine Co., Ltd., 59 East Huangcheng Road, Xinchang, Zhejiang 312500, P. R. China
 Shanghai Laiyi Center for Biopharmaceuticals R&D, 5B, Building 8 200 Niudun Road, Pudong District, Shanghai, 201203, P. R. China
§ Key Laboratory of Biomass Chemical Engineering of Ministry of Education, Zhejiang University, 38 Zhejiang University Road, Xihu District, Hangzhou, 310007, P. R. China
Org. Process Res. Dev., Article ASAP
DOI: 10.1021/acs.oprd.7b00290
*E-mail:xcwuyuanzhang@163.com (G.-f.W.)., *E-mail:xzchen@zju.edu.cn (X.-z.C.).
Abstract Image

An efficient synthesis of avibactam starting from commercially available ethyl-5-hydroxypicolinate was completed in 10 steps and 23.9% overall yield. The synthesis features a novel lipase-catalyzed resolution, in the preparation of (2S,5S)-5-hydroxypiperidine-2-carboxylate acid, which is a valuable precusor of the key intermediate ethyl (2S,5R)-5-((benzyloxy)amino)piperidine-2-carboxylate. An optimized one-pot debenzylation/sulfation reaction, followed by cation exchange, gave the avibactam sodium salt on a 400.0 g scale.

Preparation of Avibactam Sodium Salt (1)

white crystalline solid 1 (395.0 g, 96.2%), mp 259.1–262.4 °C (decomposition);
[α]D20 = −46.40 (c = 0.79, MeOH/H2O = 1/1);
1H NMR (500 MHz, D2O) δ 4.15 (dd, J = 5.8, 2.8 Hz, 1H), 4.01 (d, J = 7.5 Hz, 1H), 3.28 (d, J = 12.2 Hz, 1H), 3.06 (d, J = 12.2 Hz, 1H), 2.23–2.09 (m, 1H), 2.06–1.96 (m, 1H), 1.94–1.82 (m, 1H), 1.81–1.69 (m, 1H).
13C NMR (126 MHz, D2O) δ 174.72 (s), 169.53 (s), 60.43 (s), 59.93 (s), 47.33 (s), 20.03 (s), 18.31 (s). IR (cm–1): 3459, 1749, 1675, 1361, 1270, 1013, 857, 768. MS (ESI) m/z: 279.0 [M + H]+.
STR1STR2


PATENT

In some embodiments, sulphaturamide or tetrabutylammonium salt of (1R,2S,5R)-7-oxo-6-(sulphooxy)-1,6-diazabicyclo[3.2.1]octane-2-carboxamide may be prepared by chiral resolution of its racemic precursor trans-7-oxo-6-(phenylmethoxy)-1,6-diazabicyclo[3.2.1]octane-2-carboxamide, the preparation of which is described in Example 33a Stage A in Application WO 02/10172. In exemplary embodiments, injection of 20 μl of a sample of 0.4 mg/mL of trans-7-oxo-6-(sulphooxy)-1,6-diazabicyclo[3.2.1]octane-2-carboxamide, eluted on a Chiralpak ADH column (5 25 cm×4.6 mm) with heptane-ethanol-diethylamine mobile phase 650/350/0.05 vol at 1 mL/min makes it possible to separate the (1R,2S,5R) and (1S,2R,5S) enantiomers with retention times of 17.4 minutes and 10.8 minutes respectively. The sulphaturamide is then obtained by conversion according to the conditions described in Example 33a Stage B then Stage C and finally in Example 33b of Application WO 02/10172.

In other embodiments, the sulphaturamide can be prepared from the mixture of the oxalate salt of (2S)-5-benzyloxyamino-piperidine-2-carboxylic acid, benzyl ester (mixture (2S,5R)/(2S,5S) ˜50/50) described in application FR2921060.

For example, the preparation may proceed in the following stages:

Figure US08835455-20140916-C00006

EXAMPLES Example 1 Preparation and characterization of amorphous trans-7-oxo-6-(sulphooxy)-1,6-diazabicyclo[3,2,1]octane-2-carboxamide sodium salt

Amorphous trans-7-oxo-6-(sulphooxy)-1,6-diazabicyclo[3,2,1]octane-2-carboxamide can be prepared as described in U.S. Pat. No. 7,112,592. The XRD pattern was obtained by mounting samples on a sample holder of Rigaku Miniflex X-ray diffractometer with the Kβ radiation of copper (λ=1.541 Å). The samples, without grinding, were put on a glass plate and were analyzed at ambient temperature and humidity. Data were collected at 0.05° interval, 2°/minute from 3°-40° 2θ. FIG. 1shows the X-ray diffraction (XRD) pattern for amorphous trans-7-oxo-6-(sulphooxy)-1,6-diazabicyclo[3,2,1]octane-2-carboxamide sodium salt.

A solution, in a water-acetone mixture (1-1), of the sodium salt of the racemic trans-7-oxo-6-(sulphoxy)-1,6-diazabicyclo[3.2.1]octane-2-carboxamide described in Example 33c of Application WO 02/10172 is evaporated under reduced pressure, under the conditions of concentration described in said example. The salt is obtained in crystallized form. The X-ray spectra (“XRPD diffraction patterns”) of the polymorphic Forms were compared. The diffraction pattern of the racemic form obtained according to the prior art is different from each of those of the polymorphic Forms.

Example 2 Preparation and characterization of Form I of trans-7-oxo-6-(sulphooxy)-1,6-diazabicyclo[3,2,1]octane-2-carboxamide sodium salt

Method I

A solution of the 5.067 g (10 mmoles) of the tetrabutylammonium salt of trans-7-oxo-6-(sulfooxy)-1,6-diazabicyclo[3,2,1]octane-2-carboxamide in 12.5 ml of 200 proof ethanol and 12.5 ml of 190 proof ethanol was filtered through a 1.6 μm filter and added to a 100 ml jacketed-reactor equipped with magnetic stirrer. The solution was warmed to an internal temperature of 35° C. Separately, a solution of 3.3 g (20 mmoles) of sodium 2-ethylhexanoate in 25 ml 200 proof ethanol was filtered through a 1.6 μm filter. 2.5 ml of this solution was added to the reactor and the mixture was stirred for 1 h at 35° C. Crystallization occurred during this time. The remainder of the sodium 2-ethylhexanoate solution was added over 20 min. The mixture was stirred for an additional 1 h at 35° C., followed by 12 h at 25° C. The mixture was cooled to 0° C. for 2 h. The crystals were isolated by filtration and washed with 10 ml ethanol. The crystals were dried under vacuum at 35° C. for 16 h. 2.72 g of the sodium salt of trans-7-oxo-6-(sulfooxy)-1,6-diazabicyclo[3,2,1]octane-2-carboxamide (Form I) was obtained, corresponding to a yield of 95%

PATENT

http://www.google.com/patents/WO2014135930A1?cl=en

Example -1

Preparation of sodium salt of (2S, 5R)-sulfuric acid mono-{2-carboxamido-7-oxo-l,6-diaza- bicyclo Γ3.2.11 octane

Step-1: Preparation of (2S, 5R)-2-Carboxamido-6-benzyloxy-7-oxo-l,6-diaza- bicyclo [3.2.1] octane:

Method-1:

The starting compound ((2S, 5R)-sodium 6-benzyloxy-7-oxo-l,6-diaza-bicyclo [3.2.1] octane-2-carboxylate; compound of Formula (II)) was prepared according to a procedure disclosed in Indian Patent Application No. 699/MUM/2013. To a 100 ml round bottom flask equipped with magnetic stirrer was charged (2S, 5R)-sodium 6-benzyloxy-7- oxo-l,6-diaza-bicyclo [3.2.1] octane-2-carboxylate (10.0 gm, 0.033 mol), followed by freshly prepared HOBt. ammonia complex (10.0 gm, 0.066 mol), EDC hydrochloride (9.62 gm, 0.050 mol) and 1-hydroxy benzotriazole (4.51 gm, 0.033 mol). To this mixture of solids, water (30 ml) was added at about 35°C, and stirring was started. Precipitation occurred after 30 minutes. The reaction mixture was stirred for additional 20 hours at about 35°C. Dichloro methane (150 ml) was added to the suspension and the reaction mass was allowed to stir for 10 minutes. The layers were separated. Aqueous layer was washed with additional dichloro methane (50 ml). Combined organic layer was evaporated under vacuum to provide a residue (21 gm). The residue was stirred with acetone (21 ml) for 30 minutes and filtered under suction to provide (2S, 5R)-2-carboxamido-6-benzyloxy-7-oxo-l,6-diaza- bicyclo [3.2.1] octane as a white solid in 5.5 gm quantity in 60% yield after drying under vacuum at about 45 °C.

Analysis

H!NMR (DMSO-de)

7.35 -7.45 (m, 6H), 7.25 (bs, 1H), 4.89 – 4.96 (dd, 2H), 3.68 (d, 1H), 3.62 (s, 1H), 2.90 (s, 2H), 2.04 – 2.07 (m, 1H), 1.70-1.83 (m, 1H), 1.61-1.66 (m, 2H).

MS (ES+) C14H17N3O3 = 276.1 (M+l) Purity: 93.95% as determined by HPLC Specific rotation: [a]25 D – 8.51° (c 0.5%, CHC13) Method-2:

Alternatively, the above compound was prepared by using the following process. To a 50 ml round bottom flask equipped with magnetic stirrer was charged a solution of (2S, 5R)- sodium 6-benzyloxy-7-oxo-l,6-diaza-bicyclo [3.2.1] octane-2-carboxylate (1 gm, 0.003 mol) in water (15 ml) followed by EDC hydrochloride (1 gm, 0.005 mol) and 1- hydroxybenzotriazole (0.39 gm, 0.003 mol) at 35°C under stirring. The reaction mass was stirred for 1 hour to obtain a white suspension. At this point, aqueous ammonia was added (2 ml, 40% w/v), under stirring. The reaction mixture was stirred for additional 5 hours. The suspension was filtered, washed with additional water (10 ml) to provide (2S, 5R)-2- carboxamido-6-benzyloxy-7-oxo-l,6-diaza-bicyclo[3.2.1] after drying under vacuum at 45°C in 0.21 gm quantity.

Step-2: Preparation of Tetrabutyl ammonium salt of (2S, 5R)-2-carboxamido-6-sulfooxy-7- oxo-l,6-diaza-bicyclo [3.2.1] octane:

To a Parr shaker bottle, was charged (2S, 5R)-2-carboxamido-6-benzyloxy-7-oxo-l,6- diaza-bicyclo [3.2.1] octane (7.0 gm, 0.025 mol) followed by a 1:1 mixture of N,N- dimethylformamide and dichloro methane (35 ml: 35 ml). To the clear solution was added 10% palladium on carbon (1.75 gm) and hydrogen pressure was applied up to 50 psi. The suspension was shaken for 3 hours at 35°C. The catalyst was removed by filtering the reaction mixture over celite bed. The catalyst bed was washed with dichloro methane (30 ml). Combined filtrate was evaporated under vacuum at a temperature below 40°C to obtain an oily residue. The oily residue (4.72 gm) was dissolved in N,N-dimethylformamide (35 ml) and to the clear solution was added sulfur trioxide.DMF complex at 10°C under stirring in one lot. The mixture was allowed to stir at 35°C for additional 2 hours. As TLC showed complete conversion, 10% aqueous solution of tetrabutyl ammonium acetate (9.44 gm, 0.031 mol, in 30 ml water) was added under stirring and the reaction mixture was stirred for overnight and then subjected to high vacuum distillation on rotavapor by not exceeding temperature above 40°C to obtain a residue. Xylene (50 ml) was added to the residue and similarly evaporated to remove traces of DMF. The dry residue thus obtained was stirred with water (70 ml) and extracted with dichloro methane (70 ml x 2). Combined organic extract was dried over sodium sulfate and solvent was evaporated under vacuum below 40°C to obtain oily residue in 7 gm quantity as a crude product. It was stirred with methyl isobutyl ketone (21 ml) for 30 minutes at about 35°C to obtain a white solid in 5.9 gm quantity as a tetrabutyl ammonium salt of (2S, 5R)-2-carboxamido-6-sulfooxy-7-oxo-l,6-diaza-bicyclo[3.2.1]octane in pure form in 46% yield.

Analysis

NMR: (CDC13)

6.63 (s, 1H), 5.48 (s, 1H), 4.34 (br s, 1H), 3.90 (d, 1H), 3.27-3.40 (m, 9H), 2.84 (d, 1H), 2.38 (dd, 1H), 2.21-2.20 (m, 1H), 1.60-1.71 (m, 12H), 1.40-1.50 (m, 8H), 1.00 (t, 12H).

MS (ES-) C7H10N3O6S. N(C4H9)4 = 264.0 (M-l) as a free sulfonic acid.

Purity: 98.98% as determined by HPLC.

Specific rotation: [a]25 D – 30.99° (c 0.5%, MeOH)

Step-3: Synthesis of Sodium salt of (2S, 5R)-2-carboxamido-6-sulfooxy-7-oxo-l,6-diaza- bicyclo [3.2.1] octane

To a 100 ml round bottom flask equipped with magnetic stirrer was charged tetrabutyl ammonium salt of (2S, 5R)-2-carboxamido-6-sulfooxy-7-oxo-l,6-diaza-bicyclo[3.2.1]octane ( 5.5 gm, 0.0108 mol) followed by ethanol (28 ml) to provide a clear solution under stirring at about 35°C. To the reaction mixture was added a solution of sodium 2-ethyl hexanoate (3.6 gm, 0.021 mol) dissolved in ethanol (28 ml) in one lot under stirring to provide precipitation. The suspension was stirred for additional 2 hours to effect complete precipitation at about 35°C. The reaction mixture was filtered under suction and the wet cake was washed with acetone (30 ml x 2). The wet cake was dried at 40°C under vacuum to provide sodium salt of (2S, 5R)-2-carboxamido-6-sulfooxy-7-oxo-l,6-diaza-bicyclo[3.2.1]octane as a white solid in 2.6 gm quantity in 83% yield.

Analysis

H!NMR (DMSO-d6)

7.39 (s, 1H), 7.24 (s, 1H), 3.98 (s, 1H), 3.68 (d, 1H), 3.02 (d, 1H), 2.92 (d, 1H), 2.00- 2.10 (m, 1H), 2.80-2.90 (m, 1H), 1.55-1.70 (m, 2H).

MS (ES-) C7H10N3O6SNa = 264.0 (M-l) as a free sulfonic acid;

Purity: 97.98% as determined by HPLC

Specific rotation: [a]25 D – 49.37° (c 0.5%, water)

Powder X-ray diffractogram: (degrees 2 theta):


PATENT

http://www.google.com/patents/WO2015150941A1?cl=en


References

  1.  “Full Prescribing Information: AVYCAZ™ (ceftazidime-avibactam) for Injection, for intravenous use”. ©2015 Actavis. All rights reserved. Retrieved 1 June 2015.
  2.  Zhanel, GG (2013). “Ceftazidime-avibactam: a novel cephalosporin/β-lactamase inhibitor combination”. Drugs 73 (2): 159-77.doi:10.1007/s40265-013-0013-7. PMID 23371303.
  3.  “Actavis Announces FDA Acceptance of the NDA Filing for Ceftazidime-Avibactam, a Qualified Infectious Disease Product”. Actavis—a global, integrated specialty pharmaceutical company—Actavis. Actavis plc. Retrieved 1 June 2015.
  4. Ehmann, DE; Jahic, H; Ross, PL; Gu, RF; Hu, J; Durand-Réville, TF; Lahiri, S; Thresher, J; Livchak, S; Gao, N; Palmer, T; Walkup, GK; Fisher, SL (2013). “Kinetics of Avibactam Inhibition against Class A, C, and D β-Lactamases”. The Journal of biological chemistry 288 (39): 27960–71. doi:10.1074/jbc.M113.485979. PMC 3784710. PMID 23913691.
  5.  “www.accessdata.fda.gov” (PDF).

External links


ChemSpider 2D Image | Avibactam | C7H11N3O6S

Patent Submitted Granted
NOVEL CRYSTALLINE FORMS OF TRANS-7-OXO-6-(SULPHOOXY)-1,6-DIAZABICYCLO[3,2,1]OCTANE-2-CARBOXAMIDE SODIUM SALT [US2014349967] 2014-08-07 2014-11-27
PROCESS FOR PREPARING A COMPOUND USEFUL FOR PRODUCING AN OPTICALLY ACTIVE DIAZABICYCLOOCTANE COMPOUND [US2014303375] 2014-05-27 2014-10-09
QUICK METHOD FOR DETECTING ENYZMES AND MICROORANISMS [US2013089883] 2011-03-01 2013-04-11
Crystalline forms of trans-7-oxo-6-(sulphooxy)-1,6-diazabicyclo[3,2,1]octane-2-carboxamide sodium salt [US8835455] 2013-05-24 2014-09-16
WO2009091856A2 * Jan 15, 2009 Jul 23, 2009 Merck & Co Inc Beta-lactamase inhibitors
WO2012086241A1 * Jun 30, 2011 Jun 28, 2012 Meiji Seika Pharma Co., Ltd. Optically-active diazabicyclooctane derivative and method for manufacturing same
INMU06992013A Title not available
US7112592 Jul 24, 2001 Sep 26, 2006 Aventis Pharma S.A. Azabicyclic compounds, preparation thereof and use as medicines, in particular as antibacterial agents
Avibactam
Avibactam structure 2.svg
Avibactam ball-and-stick model.png
Systematic (IUPAC) name
[(2S,5R)-2-Carbamoyl-7-oxo-1,6-diazabicyclo[3.2.1]octan-6-yl] hydrogen sulfate
Clinical data
Trade names Avycaz (formulated with ceftazidime)
Legal status
Routes of
administration
intravenous
Pharmacokinetic data
Bioavailability 100% (intravenous)
Protein binding 5.7–8.2%[1]
Metabolism nil
Onset of action increases in proportion to dose
Excretion Renal (97%)
Identifiers
CAS Number 1192500-31-4
ATC code J01
PubChem CID: 9835049
ChemSpider 8010770
ChEBI CHEBI:85984 Yes
ChEMBL CHEMBL1689063
Chemical data
Formula C7H11N3O6S
Molecular mass 265.24 g/mol

SEE BACTAM SERIES…………..http://apisynthesisint.blogspot.in/p/bactam-series.html


////////

[Na+].NC(=O)[C@@H]2CC[C@@H]1CN2C(=O)N1OS([O-])(=O)=O

C1CC(N2CC1N(C2=O)OS(=O)(=O)O)C(=O)N

update…………

Avibactam, a β-lactamase inhibitor,in combination with Ceftazidime (Zavicefta) has recently been approved by EMA for treatment of complicated intra-abdominal infections (cIAI), complicated urinary tract infections (cUTI), and hospital-acquired pneumonia (HAP), including ventilator-associated pneumonia (VAP). The EMA also approved Zavicefta for the treatment of infections caused by aerobic Gram-negative organisms in adult patients who have limited treatment options.
Avibactam was originally developed by the Aventis infection division in Romainville (France), which later became Novexel. During the Phase II studies, however, commercial developments led to the project becoming a co-development between AstraZeneca and Forest Laboratories.

Development of a Manufacturing Route to Avibactam, a β-Lactamase Inhibitor

Pharmaceutical Technology and Development, AstraZeneca, Silk Road Business Park, Macclesfield SK10 2NA, United Kingdom
Chemical Development, Forest Laboratories Inc., 45 Adams Avenue, Hauppauge, New York 1178, United States
Org. Process Res. Dev., Article ASAP, http://pubs.acs.org/doi/full/10.1021/acs.oprd.6b00268
DOI: 10.1021/acs.oprd.6b00268

Abstract

Abstract Image

Process development work to provide an efficient, robust, and cost-effective manufacturing route to avibactam, a β-lactamase inhibitor is presented herewith. Aspects of this optimization work include the counterintuitive introduction of a protecting group to effect a difficult urea formation and the use of controlled feed hydrogenation conditions to facilitate an elegant one pot debenzylation and sulfation reaction. Overall, the commercial process delivers avibactam in much improved yield with significant reduction in the environmental footprint.

Preparation of Benzyl (2S,5R)-5-[(Benzyloxy)amino]piperidine-2-carboxylate Ethanedioate (1:1)

1H NMR (400 MHz, DMSO) δ: 1.52 (1H, m), 1.70 (1H, m), 1.94 (1H, d, J = 12.3 Hz), 2.22 (1H, dd, J = 13.8 Hz, J = 3.6 Hz), 2.79 (1H, t, J = 11.5 Hz), 3.27 (1H, m), 3.46 (1H, d, J = 11.5 Hz), 4.14 (1H, dd, J = 12.3 Hz, J = 3.2 Hz), 4.68 (2H, s), 5.24 (2H, s), 7.34 (10H, m). 13C NMR (100 MHz, DMSO) δH 25.4 (s), 26.1 (s), 46.5 (s), 54.0 (s), 56.4 (s), 67.3 (s), 76.4 (s), 128.5 (m), 135.7 (s), 138.5 (s), 164.7 (s), 167.5 (s). HRMS Calcd for C20H25N2O3: 341.1860; HRMS found [M+H]+: 341.1858.

Preparation of (2S,5R)-5-[(Benzyloxy)amino]piperidine-2-carboxamide

1H NMR (400 MHz, DMSO) δH 1.12 (1H, m), 1.27 (1H, m), 1.83 (2H, m), 2.22 (1H, dd, J = 10.1 Hz, J = 12.0 Hz), 2.76 (1H, m), 2.89 (1H, dd, J = 2.8 Hz, J = 10.9 Hz), 3.14 (1H, dd, J = 4.1 Hz, J = 12.0 Hz), 4.58 (2H, s), 6.46 (1H, d, J = 5.6 Hz), 6.91 (1H, s), 7.09 (1H, s), 7.32 (5H, m). 13C NMR (100 MHz, DMSO) δH 28.4 (s), 29.2 (s), 49.5 (s), 57.5 (s), 59.8 (s), 76.3 (s), 128.3 (m), 138.9 (s), 175.6 (s). HRMS Calcd for C13H20N3O2: 250.1550; HRMS found [M+H]+: 250.1551.

Preparation of (2S,5R)-6-(Benzyloxy)-7-oxo-1,6-diazabicyclo[3.2.1]octane-2-carboxamide

1H NMR (400 MHz, DMSO) δH 1.65 (2H, m), 1.83 (1H, m), 2.07 (1H, m), 2.91 (2H, s), 3.63 (1H, s), 3.69 (1H, d, J = 6.7 Hz), 4.92 (2H, dd, J = 18.1 Hz, J = 11.4 Hz), 7.38 (7H, m). 13C NMR (100 MHz, DMSO) δH 18.6 (s), 21.1 (s), 47.2 (s), 57.5 (s), 60.1 (s), 77.4 (s), 129.0 (m), 136.3 (s), 167.9 (s), 171.8 (s). HRMS Calcd for C14H18N3O3: 276.1343; HRMS found [M+H]+: 276.1336

Preparation of Tetrabutylammonium [(2S,5R)-2-Carbamoyl-7-oxo-1,6-diazabicyclo[3.2.1]octan-6-yl] Sulfate

1H NMR (400 MHz, CDCl3) δH 1.00 (12H, t, J = 7.2 Hz), 1.45 (8H, m), 1.67 (9H, m), 1.87 (1H, m), 2.16 (1H, m), 2.37 (1H, dd, J = 15.0 Hz, J = 7.0 Hz), 2.87 (1H, d, J = 11.6 Hz), 3.31 (9H, m), 3.91 (1H, d, J = 7.9 Hz), 4.33 (1H, s), 5.87 (1H, s), 6.69 (1H, s). 13C NMR (100 MHz, D2O) δH 12.8 (s), 18.1 (s), 19.1 (s), 19.9 (s), 23.1 (s), 47.2 (s), 58.1 (s), 59.8 (s), 60.3 (s), 169.4 (s), 174.7 (s). HRMS Calcd for C7H10N3O6S: 264.0296; HRMS found [M–H]–: 264.0298

AVIBACTAM

1H NMR (500 MHz, DMSO) δH 1.63 (2H, m), 1.83 (1H, m), 2.05 (1H, m), 2.90 (1H, d, J = 11.8 Hz), 3.00 (1H, d, J = 11.6 Hz), 3.67 (1H, d, J = 6.9 Hz), 3.98 (1H, s), 7.29 (1H, s), 7.44 (1H, s). 13C NMR (100 MHz, D2O) δH 18.1 (s), 19.9 (s), 47.2 (s), 59.8 (s), 60.3 (s), 169.4 (s), 174.7 (s). HRMS Calcd for C7H12N3O6S: 266.0441; HRMS found [M+H]+: 266.0441


Open Babel bond-line chemical structure with annotated hydrogens.<br>Click to toggle size.

<sup>1</sup>H NMR spectrum of C<sub>7</sub>H<sub>10</sub>N<sub>3</sub>O<sub>6</sub>S<sub></sub> in CDCL3 at 400 MHz.<br>Click to toggle size.

Shifts

Index Name Shift (ppm)
16 H6 4.573
27 H5 2.416
18 H7 1.742
19 H8 1.870
26 H4 1.949
11 H2 3.497
12 H3 4.412
14 H1 5.267

///////////