


| Patent ID | Date | Patent Title |
|---|---|---|
| US2012077812 | 2012-03-29 | BICYCLIC COMPOUNDS AND USE AS ANTIDIABETICS |
| US8101634 | 2012-01-24 | BICYCLIC COMPOUNDS AND USE AS ANTIDIABETICS |
PROUD Indian WORLD RECORD VIEWS holder on THIS BLOG, ………live, by DR ANTHONY MELVIN CRASTO, Worldpeaceambassador, Worlddrugtracker, Helping millions, 100 million hits on google, pushing boundaries,2.5 lakh plus connections worldwide, 45 lakh plus VIEWS on this blog in 227 countries, 7 CONTINENTS ……A 90 % paralysed man in action for you, I am suffering from transverse mylitis and bound to a wheel chair, [THIS BLOG HOLDS WORLD RECORD VIEWS ] A PROUD INDIAN
Home » Uncategorized (Page 69)
 DRUG APPROVALS BY DR ANTHONY MELVIN CRASTO
.....FOR BLOG HOME CLICK HERE


 Subscribe in a reader
Enter your email address:
Delivered by FeedBurner
Subscribe to New Drug Approvals by Email


 | shivkr2 on SPSR Excellence Award 2025 – S… |
 | Bonkasaurus on Infigratinib phosphate |
 | PALOVAROTENE | ORGAN… on Palovarotene |
| Pfizer just purchase… on Temanogrel | |
| Pfizer To Cash In On… on Temanogrel |

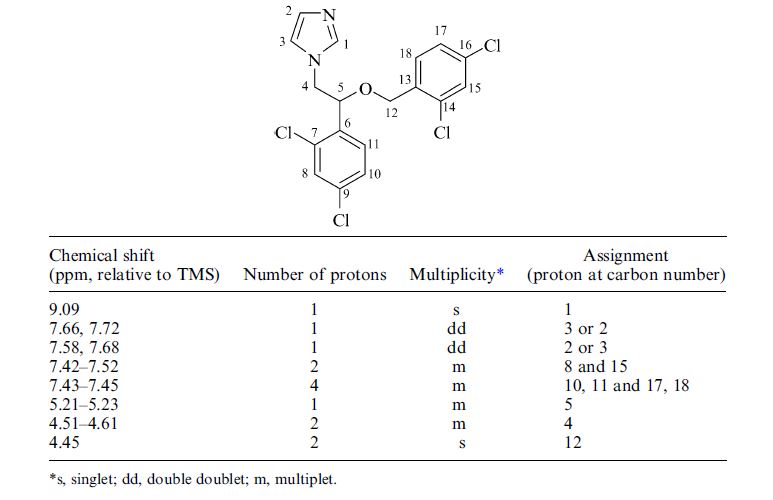
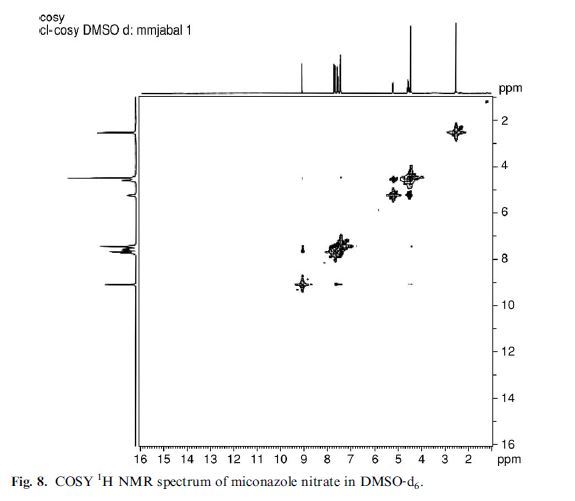
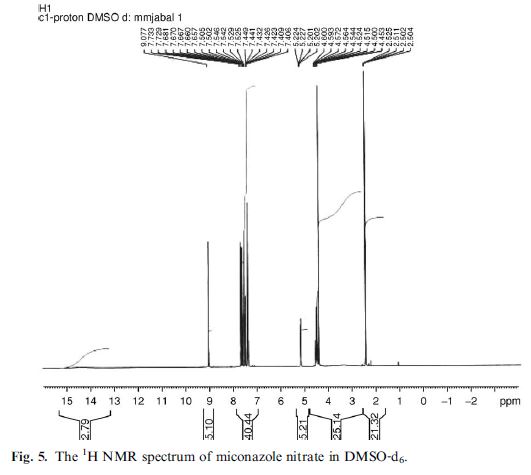
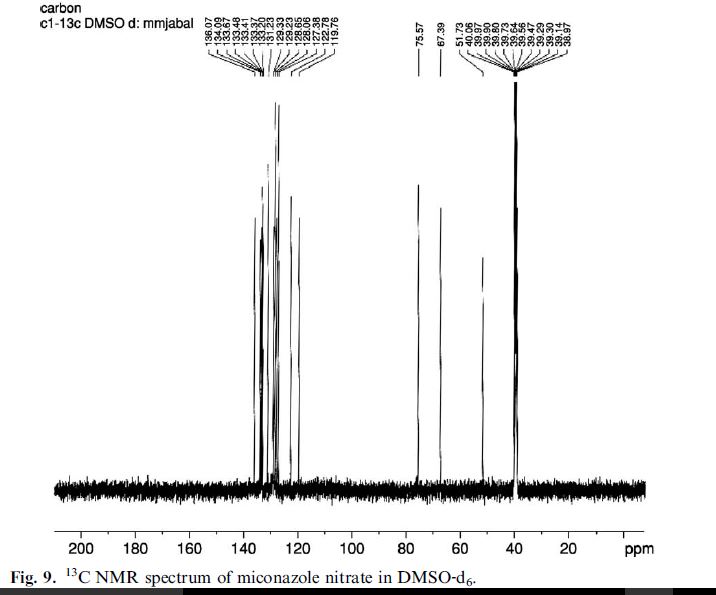
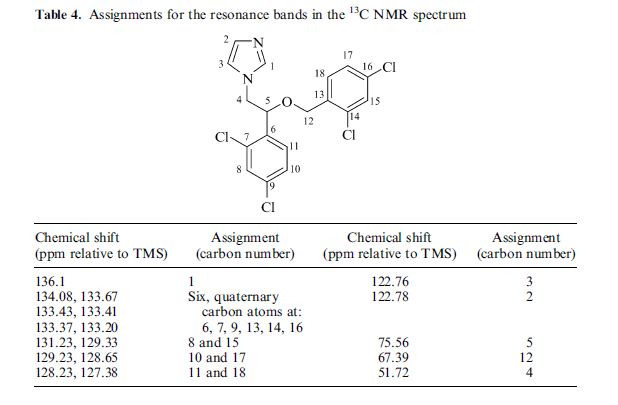
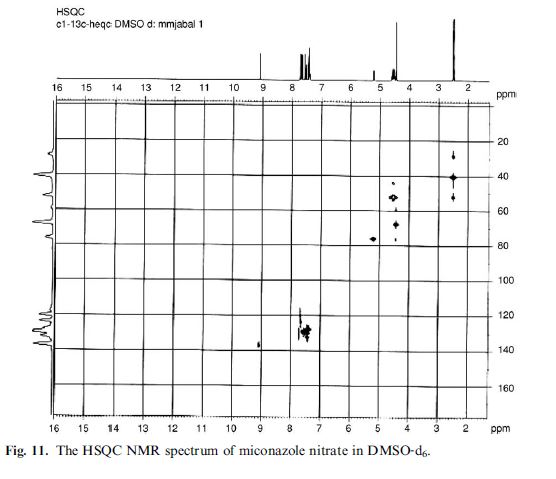
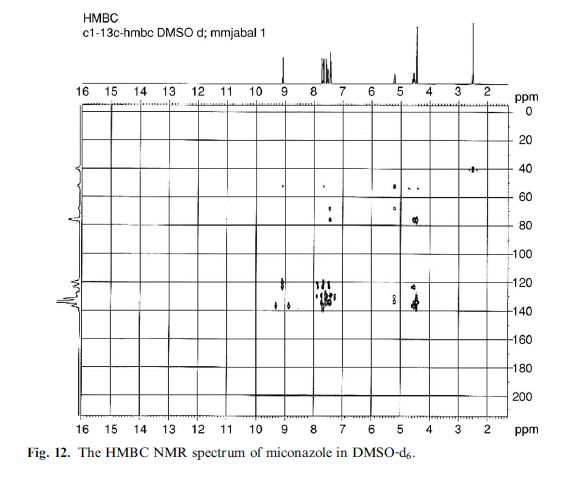
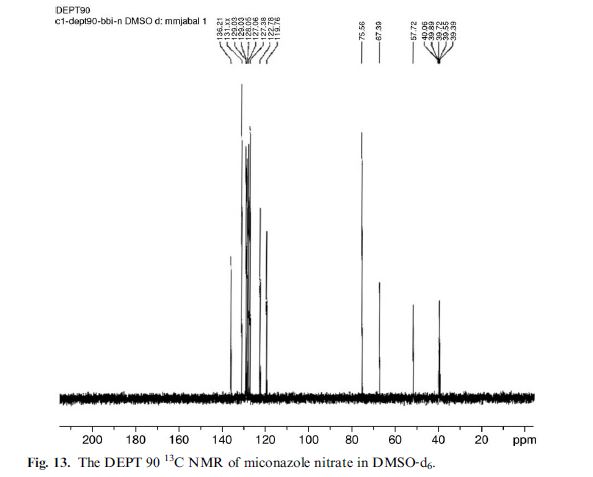
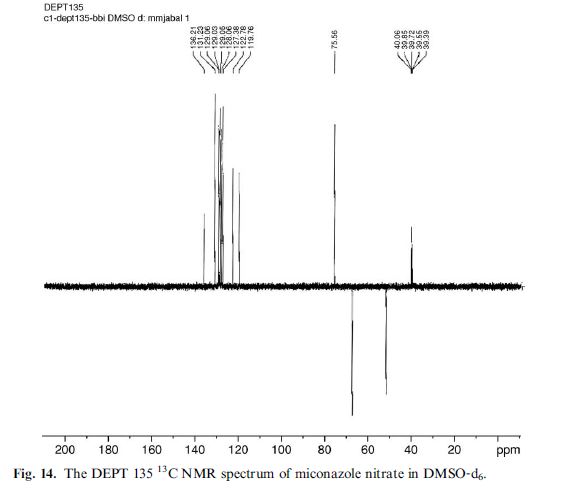

 click on above image for clear view
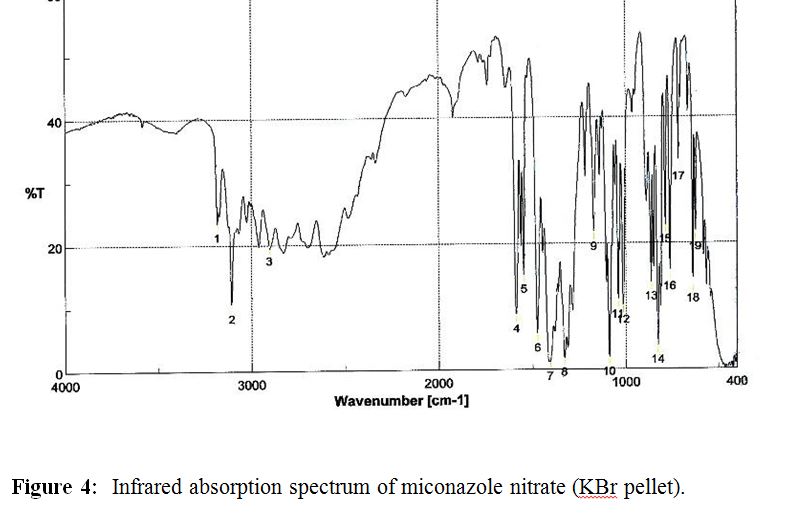
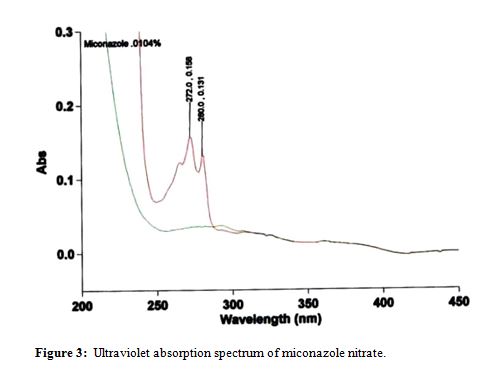
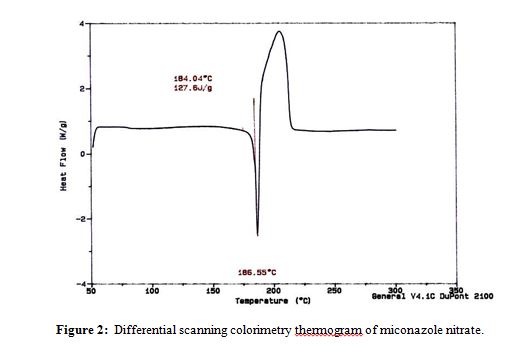
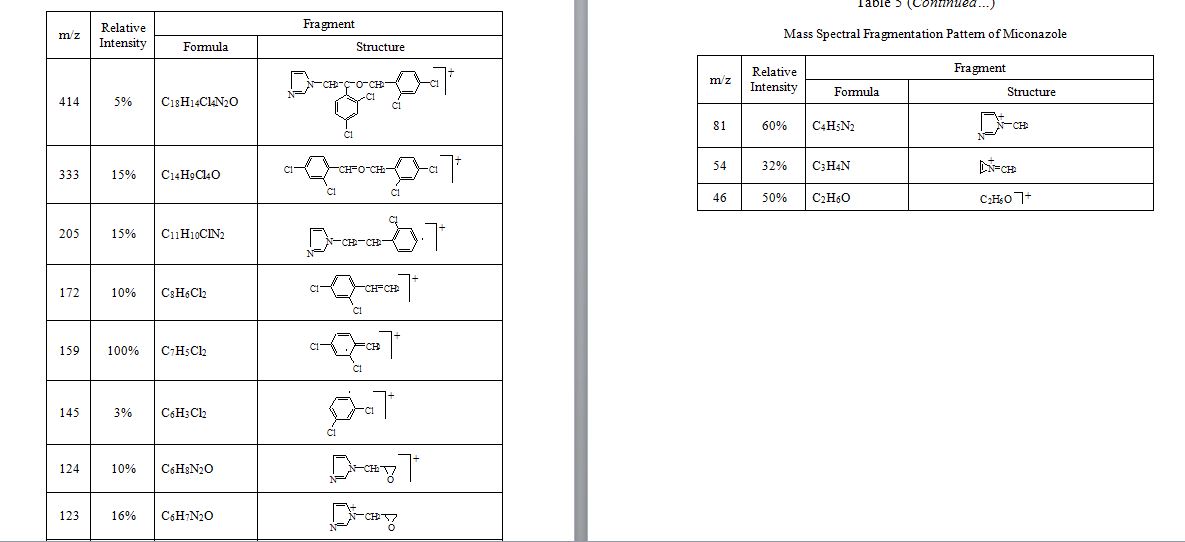
MORE GRAPHS
13C


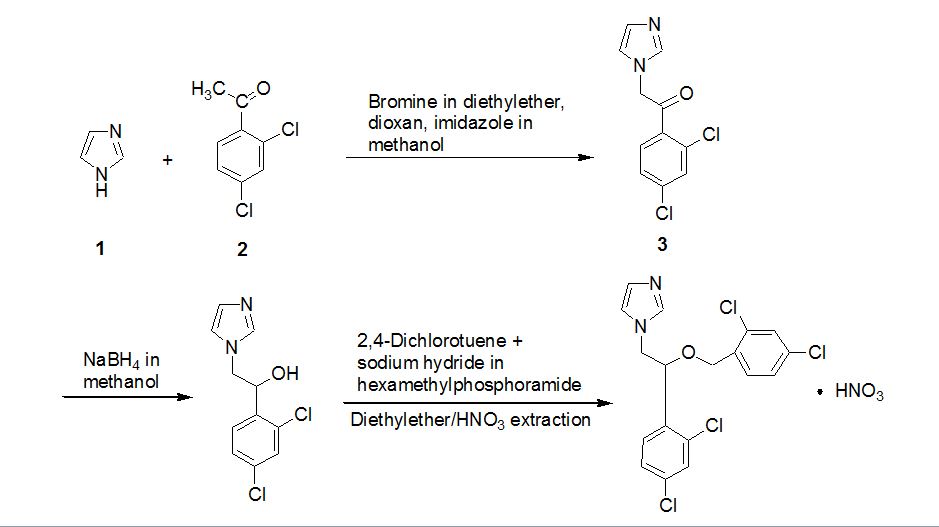
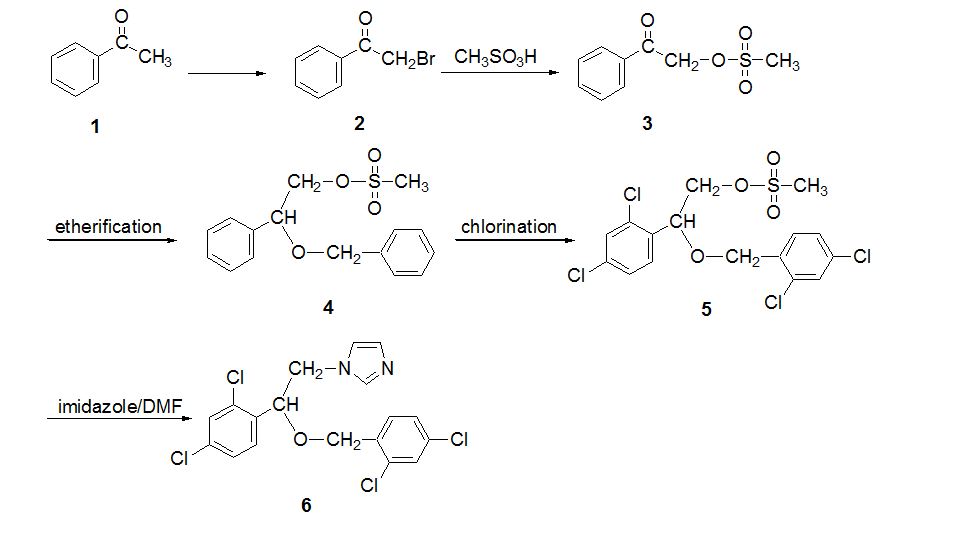
Miconazole is an imidazole antifungal agent, developed by Janssen Pharmaceutica, commonly applied topically to the skin or tomucous membranes to cure fungal infections. It works by inhibiting the synthesis of ergosterol, a critical component of fungal cell membranes. It can also be used against certain species of Leishmania protozoa which are a type of unicellular parasites that also contain ergosterol in their cell membranes. In addition to its antifungal and antiparasitic actions, it also has some antibacterialproperties. It is marketed in various formulations under various brand names.


| Salt | ATC | Formula | MM | CAS |
|---|---|---|---|---|
| – | A01AB09 A07AC01 D01AC02 G01AF04 J02AB01 S02AA13 |
C 18 H 14 Cl 4 N 2 O | 416.14 g / mol | 22916-47-8 |
| mononitrate | A01AB09 A07AC01 D01AC02 G01AF04 J02AB01 S02AA13 |
C 18 H 14 Cl 4 N 2 O ⋅ HNO 3 | 479.15 g / mol | 22832-87-7 |
| Synthesis of a) |
|---|
  |
| A country | Tradename | Manufacturer |
|---|---|---|
| Germany | Castellani | Hollborn |
| Daktar | McNeil | |
| Derma-Mikotral | Rosen Pharma | |
| Fungur | HEXAL | |
| Gyno-Daktar | Janssen-Cilag, 1974 | |
| Gyno-Mikotral | Rosen Pharma | |
| Infektozoor Mundgel | Infectopharm | |
| Mikobeta | betapharm | |
| Mikotar | Dermapharm | |
| Mikoderm | Engelhard | |
| Mikotin | Ardeypharm | |
| Vobamik | Almirall Hermal | |
| France | Daktapin | Janssen-Cilag |
| Gyno-Daktapin | Janssen-Cilag | |
| Loramik | Bioalliance | |
| United Kingdom | Gyno-Daktapin | Janssen-Cilag |
| Italy | Daktapin | Janssen-Cilag |
| Mikonal | Ecobi | |
| Mikotef | LPB | |
| Miderm | Mendelejeff | |
| Nizakol | PS Pharma | |
| Pivanazolo | Medestea | |
| Prilagin | Sofar | |
| Japan | Florid | Mochida |
| USA | Fungoid | Pedinol |
| Ukraine | GІNEZOL 7 | Sagmel, Іnk., USA |
| MІKONAZOL-Darnitsa | CJSC “Farmatsevtichna FIRMA” Darnitsa “, m. Kyiv, Ukraine | |
| MІKOGEL | BAT “Kiїvmedpreparat”, m. Kyiv, Ukraine | |
| various generic drugs | ||

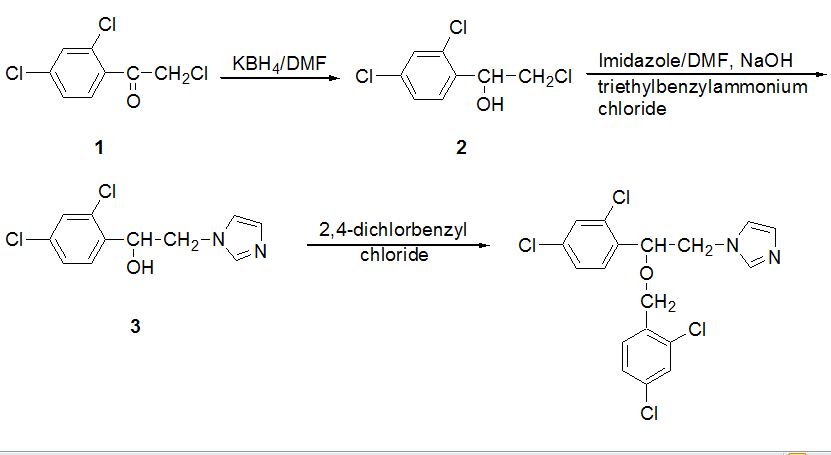
 Title: Miconazole
 |
|
 |
|
| Systematic (IUPAC) name | |
|---|---|
|
(RS)-1-(2-(2,4-Dichlorobenzyloxy)-2-(2,4-dichlorophenyl)ethyl)-1H-imidazole
|
|
| Clinical data | |
| Trade names | Desenex, Monistat, Zeasorb-AF |
| AHFS/Drugs.com | Monograph |
| MedlinePlus | a601203 |
| Pregnancy category |
|
| Routes of administration |
topical, vaginal, sublabial,oral |
| Legal status | |
| Legal status |
|
| Pharmacokinetic data | |
| Bioavailability | n/a |
| Metabolism | n/a |
| Biological half-life | n/a |
| Excretion | n/a |
| Identifiers | |
| CAS Number | 22916-47-8  |
| ATC code | A01AB09 (WHO)A07AC01 (WHO)D01AC02 (WHO)G01AF04 (WHO)J02AB01 (WHO)S02AA13 (WHO) |
| PubChem | CID 4189 |
| IUPHAR/BPS | 2449 |
| DrugBank | DB01110 |
| ChemSpider | 4044 |
| UNII | 7NNO0D7S5M |
| KEGG | D00416 |
| ChEBI | CHEBI:6923 |
| ChEMBL | CHEMBL91 |
| Chemical data | |
| Formula | C18H14Cl4N2O |
| Molar mass | 416.127 g/mol |
| Chirality | Racemic mixture |

 Googleplus
 amcrasto@gmail.com
///////////


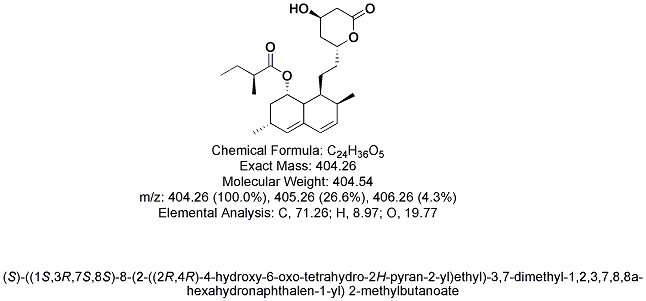
Lovastatin (Merck’s Mevacor) is a statin drug, used for lowering cholesterol (hypolipidemic agent) in those withhypercholesterolemia to reduce risk of cardiovascular disease. Lovastatin is a naturally occurring compound found in food such asoyster mushrooms,[2] red yeast rice,[3] and Pu-erh.[4]
The primary uses of lovastatin is for the treatment of dyslipidemia and the prevention of cardiovascular disease.[5] It is recommended to be used only after other measures, such as diet, exercise, and weight reduction, have not improved cholesterol levels.[5]

Pleurotus ostreatus, the oyster mushroom, naturally contains up to 2.8% lovastatin on a dry weight basis.[15]
Structure


History
Compactin and lovastatin, natural products with a powerful inhibitory effect on HMG-CoA reductase, were discovered in the 1970s, and taken into clinical development as potential drugs for lowering LDL cholesterol.
However, in 1980, trials with compactin were suspended for undisclosed reasons (rumoured to be related to serious animal toxicity). Because of the close structural similarity between compactin and lovastatin, clinical studies with lovastatin were also suspended, and additional animal safety studies initiated.
In 1982 some small-scale clinical investigations of lovastatin, a polyketide derived natural product isolated from Aspergillus terrus, in very high-risk patients were undertaken, in which dramatic reductions in LDL cholesterol were observed, with very few adverse effects. After the additional animal safety studies with lovastatin revealed no toxicity of the type thought to be associated with compactin, clinical studies resumed.
Large-scale trials confirmed the effectiveness of lovastatin. Observed tolerability continued to be excellent, and lovastatin was approved by the US FDA in 1987.
Lovastatin at its maximal recommended dose of 80 mg daily produced a mean reduction in LDL cholesterol of 40%, a far greater reduction than could be obtained with any of the treatments available at the time. Equally important, the drug produced very few adverse effects, was easy for patients to take, and so was rapidly accepted by prescribers and patients. The only important adverse effect is myopathy/rhabdomyolysis. This is rare and occurs with all HMG-CoA reductase inhibitors.
Mechanism of action
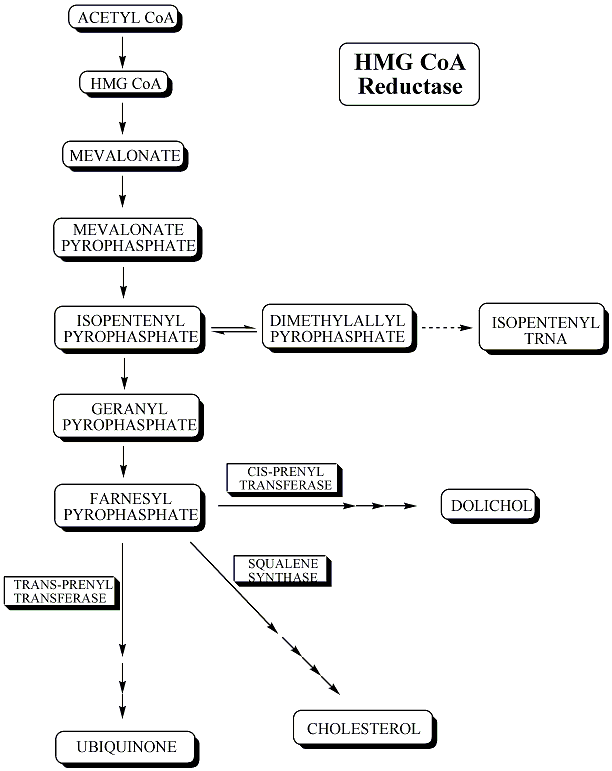
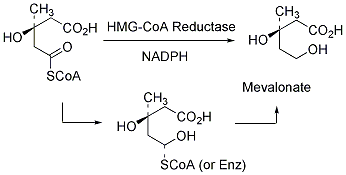
Lovastatin is an inhibitor of 3-hydroxy-3methylglutaryl-coenzyme A reductase (HMG-CoA reductase), an enzyme which catalyzes the conversion of HMG-CoA to mevalonate. Mevalonate is a required building block for cholesterol biosynthesis and lovastatin interferes with its production by acting as a competitive inhibitor for HMG-CoA which binds to the HMG-CoA reductase. Lovastatin, being inactive in the native form, the form in which it is administered, is hydrolysed to the β-hydroxy acid form in the body and it is this form which is active. Presumably, the reductase acts on the hydrolyzed lovastatin to reduce the carboxylic acid moiety.
Discovery, Biochemistry and Biology
It is now generally accepted that a major risk factor for the development of coronary heart disease is an elevated concentration of plasma cholesterol, especially lowdensity lipoprotein (LDL) cholesterol. The objective is to decrease excess levels of cholesterol to an amount consistent with maintainence of normal body function. Cholesterol is biosynthesized in a series of more than 25 separate enzymatic reactions that initially involves 3 successive condensations of acetyl-CoA units to form a 6-carbon compound, 3-hydroxy-3-methylglutaryl coenzyme A (HMG CoA). This is reduced to mevalonate and then converted in a series of reactions to the isoprenes that are building blocks of squalene, the immediate precursor to sterols, which cyclizes to lanosterol (a methylated sterol) and further metabolized to cholesterol. A number of early attempts to block the synthesis of cholesterol resulted in agents that inhibited late in the biosynthetic pathway between lanosterol and cholesterol. A major rate limiting step in the pathway is at the level of the microsomal enzyme which catalyzes the conversion of HMG CoA to mevalonic acd and which has been considered to be a prime target for pharmacologic intervention for several years.
HMG CoA reductase occurs early in the biosynthetic pathway and is among the first commited steps to cholesterol formulation. Inhibition of this enzyme could lead to accumulation of HMG CoA, a water-soluble intermediate that is then capable of being readily metabolized that is then capable of being readily metabolized to simpler molecules. This inhibition of reductase would nto lead to accumulation of lipophylic intermediates having a formal sterol ring.
Lovastatin is the first specific inhibitor of HMG CoA reductase to receive approval for the treatment of hypercholesterolemia. The first breakthrough in efforts to find a potent, specific, competitive inhibitor of HMG CoA reductase occurred in 1976 when Endo et al reported discovery of mevastatin, a highly functionalized fungal metabolite, isolated from cultures of Penicillium citrium. Mevastatin was demonstrated to be an unusually potent inhibitor of the target enzyme and of cholesterol biosynthesis. Subsequent to the first reports describing mevastatin, efforts were initiated to search for other naturally occurring inhibitors oh HMG CoA reductase. This led to the discovery of a novel fungal metabolite – Lovastatin. The structure of Lovastatin was determined to be different from that of mevastatin by the presence of a 6 alphamethyl group in the hexahydronaphthalene ring.
Key points from the study of the Biosynthesis of Lovastatin :-
– Lovastatin is comprised of 2 polyketide chains derived from acetate that are 8- and 4-
carbons long coupled in head to tail fashion.
– 6 alphamethyl group and the methyl group on the 4-carbon side chain are derived from
the methyl group of methionine, and
– 6 alphamethyl group is added before closure of the rings.
This implies that lovastatin is a unique compound synthesized by A. terreus and that mevastatin is not an intermediate in its fornmation.
Biosynthesis — Diels-Alder Catalyzed Cyclization
In vitro formation of a triketide lactone using a genetically-modified protein derived from 6-deoxyerythronolide B synthase has been demonstrated. The stereochemistry of the molecule supports the intriguing idea that an enzyme-catalyzed Diels-Alder reaction may occur during assembly of the polyketide chain. It thus appears that biological Diels-Alder reactions may be triggered by generation of reactive triene systems on an enzyme surface.


Biosynthesis – Using Broadly specific Acyltransferase
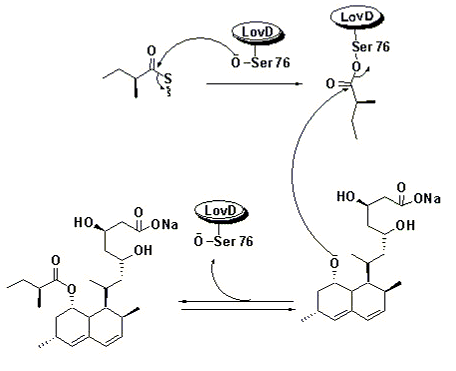
It has been found that a dedicated acyltransferase, LovD, is encoded in the lovastatin biosynthetic pathway. LovD has a broad substrate specificity towards the acyl carrier, the acyl substrate and the decalin acyl acceptor. It efficiently catalyzes the acyl transfer from coenzyme A thoesters or N-acetylcysteamine (SNAC) thioesters to monacolin J.
The biosynthesis of Lovastatin is coordinated by two iterative type I polyketide synthases and numerous accessory enzymes. Nonketide, the intermediate biosynthetic precursor of Lovastatin, is assembeled by the upstream megasynthase LovB (also known as lovastatin nonaketide synthase), enoylreductase LovC, and CYP450 oxygenases. The five carbon unit side chain is synthesized by LovF (also known as lovastatin diketide synthase) through a single condensation diketide undergoes methylation and reductive tailoring by the individual LovF catalytic domains to yield an α-S-methylbutyryl thioester covalently attached to the phosphopantetheine arm on the acyl carrier protein (ACP) domain of LovF. Encoded in the gene cluster is a 46kDa protein, LovD, which was initially identified as an esterase homolog. LovD, which was initially identified as an esterase homolog. LovD was suggested to catalyze the last step of lovastatin biosynthesis that regioselectively transacylates the acyl group from LovF to the C8 hydroxyl group of the Nonaketide to yield Lovastatin.
K. Auclair, A. Sutherland, J. Kennedy, D. J. Witter, J. P. Van den Heever, C. R. Hutchinson and J. C. Vederas, Lovastatin Nonaketide Synthase Catalyses An Intramolecular Diels-Alder Reaction Of A Substrate Analogue, J. Am. Chem. Soc., 2000, 122, 11519-11520. DOI: 10.1021/ja003216+


http://pubs.rsc.org/en/content/articlelanding/2013/np/c2np20069d/unauth#!divAbstract

http://www.hindawi.com/journals/bmri/2012/196264/#B30


Patent
https://www.google.com/patents/US6307066

PATENT
https://www.google.com/patents/WO2002009697A1?cl=en


https://www.google.com/patents/EP0625208B1?cl=en
Total Synthesis
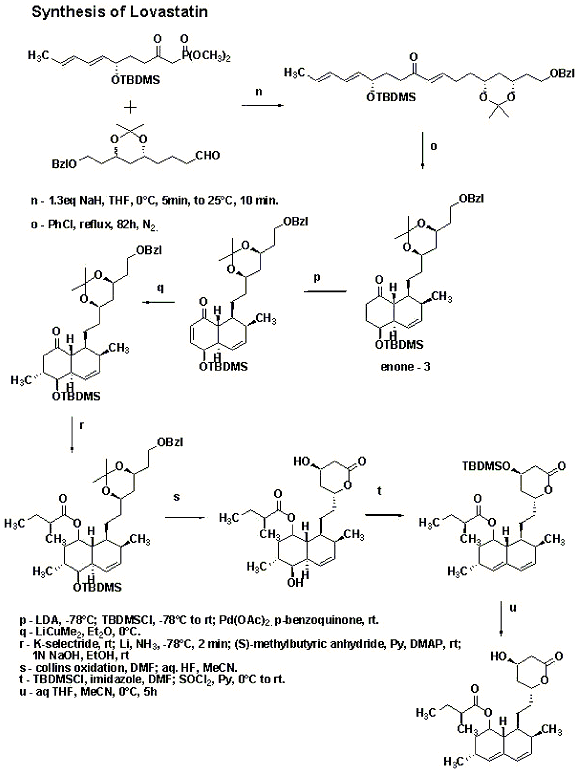
A major bulk of work in the synthesis of Lovastatin was done by M. Hirama in the 1980’s. Hirama synthesized Compactin and used one of the intermediates to follow a different path to get to Lovastatin. The synthetic sequence is shown in the schemes below. The γ-lactone was synthesized using Yamada methodology starting with aspartic acid. Lactone opening was done using lithium methoxide in methanol and then silylation to give a separable mixture of the starting lactone and the silyl ether. The silyl ether on hydrogenolysis followed by Collins oxidation gave the aldehyde. Stereoselective preparation of (E,E)-diene was accomplished by addition of trans-crotyl phenyl sulfone anion, followed by quenching with Ac2O and subsequent reductive elimination of sulfone acetate. Condensation of this with Lithium anion of dimethyl methylphosphonate gave compound 1.Compound 2 was synthesized as shown in the scheme in the synthetic procedure. Compounds 1 and 2 were then combined together using 1.3eq sodium hydride in THF followed by reflux in chlorobenzene for 82 hrs under nitrogen to get the enone 3.
Simple organic reactions were used to get to Lovastatin as shown in the scheme.

Pharmacopoeia Information
Lovastatin tablets are preserved in well closed, light resistant containers. Protected from light and stored either in a cool place or at controlled room temperature.
Lovastatin tablets are tested for Dissolution and Assay as per the USP.
Limit for Dissolution – Not less than 80% (Q) of the labeled amount of Lovastatin is dissolved in 30 mins.
Limit for Assay – Each tablet contains not less than 90% and not more than 110% of the labeled amount of Lovastatin, tested by HPLC analysis.
Lovastatin raw material contains 5 impurities – A, B, C, D and E (as shown below).

Market brands and other analogues
There are other derivatives of Lovastatin which possess cholesterol reducing activity. Simvastatin (Zocor®) is another statin closely related to Lovastatin, differing only by the presence of a methyl group in the butanoyl ester moiety. Both effective in lowering total cholesterol.
Another statin having vastly different structure but a popular drug – Atorvastatin (Lipitor®), administered as a calcium salt is a pyrrole derivative and a synthetic compound rather than a natural product.
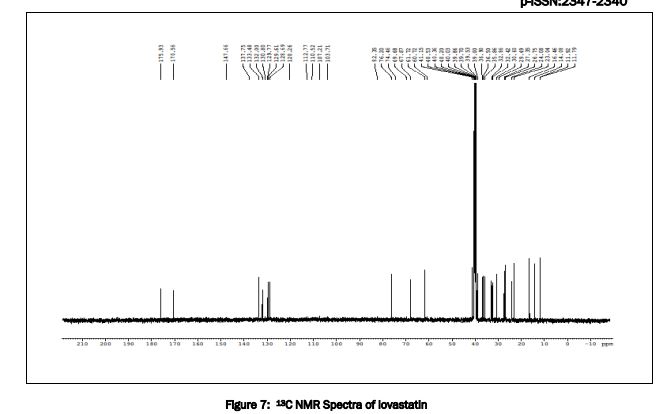
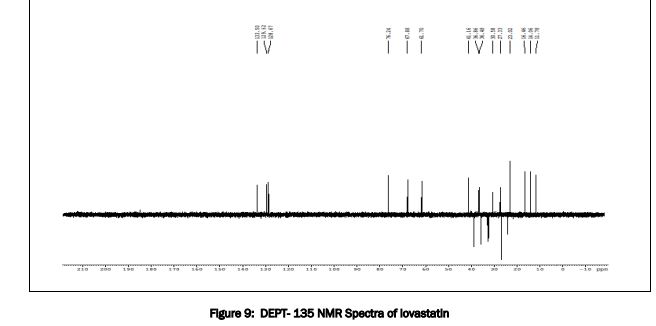
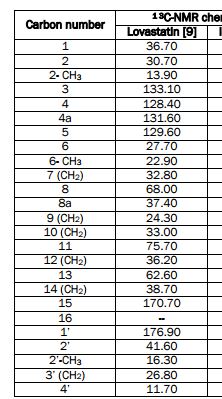
NMR

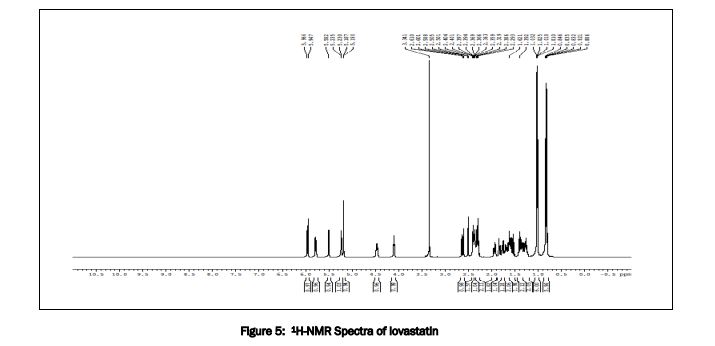
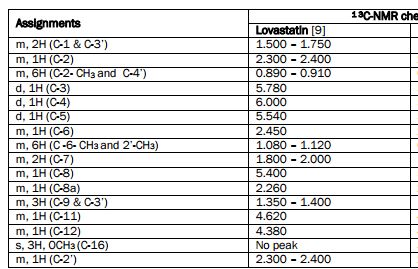
1 H NMR spectrum of lovastatin, 300 MHz, solvent CDCl 3 .
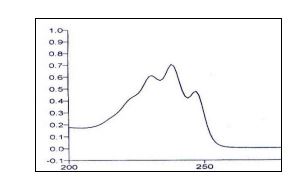 UV LOVASTATIN

Figure 6. The mean FT-IR spectra (the calibration set) and variables selected after application of UVE-PLS for modelling lovastatin (triangles) and wavenumbers for characteristic peaks for lovastatin IR spectrum (dots).
PATENT
https://www.google.com/patents/EP0702679B1?cl=en
Lovastatin is produced as a secondary metabolite of the fungusAspergillus terreus (US 4,231,938) deposited in American Type Culture Collection under Nos. ATCC 20541, ATCC 20542, and Monascus ruberdeposited in Fermentation Research Institute Agency of Industrial Science and Technology, Ministry of International Trade and Industry, Japan (DE 30 06 216 A1) under No. Ferm 4822. Other kinds of microorganisms producing lovastatin are known as well, e.g. a mutant of the microorganism Aspergillus terreus andAspergillus oryzae marked ATCC 74135.
Lovastatin is chemically 1′,2′,6′,7′,8a’-hexahydro-3,5-dihydroxy-2′,6′-dimethyl-8′-2″-methyl-1″-oxobutoxy)-1-naphtalene heptanoic acid-5-lactone (Stubbs et al., 1986) of the formula (EP 0 033 537 A1)

An active form of lovastatin is also an acid, which is chemically 1,2,6,8,8a-hexahydro-β,δ-dihydroxy-1-naphtalene heptanoic acid (Alberts et al., 1980) of the formula (EP 0 022 478 A1)

The lactone form of lovastatin is used as an agent for reducing cholesterol level in blood (Scott M.G. and Vega G.L, 1985). It inhibits the biosynthesis of mevalonic acid by inhibition of 3-hydroxy-3-methylglutaryl A reductase coenzyme (HMG-CoA reductase, E.C. 1.1.1.34) (Zubay et al., 1984).
Prior Art
After the completed fermentation, lovastatin is present in the broth in the lactone form (compound I) and in the acid form (compound II). In the isolation process as disclosed in EP 0 033 536 A2, lovastatin is extracted from the broth with ethyl acetate. The extract is concentrated by vacuum distillation. Since lovastatin is present in the lactone form as well as in the acid form and only the lactone is of commercial interest, the acid form should be converted into the lactone. The lactonisation is carried out by the reflux of the concentrate in toluene at 106 °C for 2 hours. After the lactonisation is complete, the solution is concentrated to a small volume. A pure substance is obtained by means of purifying the concentrate on columns packed with silica gel, in the presence of solvents such as ethyl acetate or n-hexane. The collected fractions are again concentrated in vacuo and then pure lovastatin crystallizes in the lactone form.
Due to the sophisticated multi-step procedure and vigorous conditions applied during the isolation, the yields of lovastatin are generally low. Different solvents, which in part exhibit toxicity, are used such as benzene, toluene, acetonitrile or ethyl acetate. Hence working with these solvents endangers the health of the persons involved and poses a problem with respect to the environment.


https://www.google.com/patents/EP0702679B1?cl=en
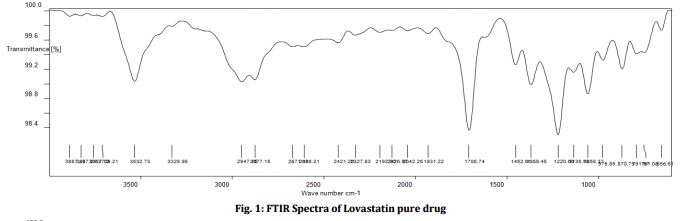
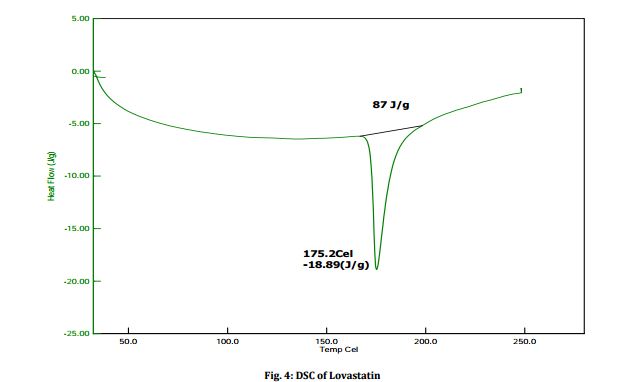
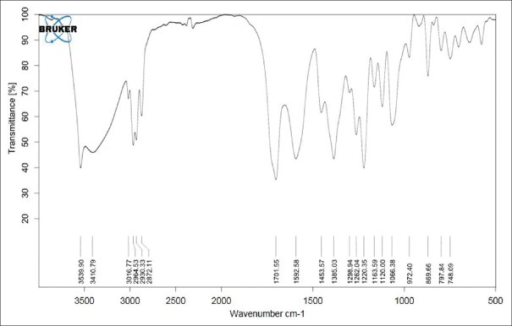 IR spectrum of lovastatin.

 .PMID 17302963.. PMID 10393901..PMID 6933445.
 |
|
 |
|
| Systematic (IUPAC) name | |
|---|---|
|
(1S,3R,7S,8S,8aR)-8-{2-[(2R,4R)-4-Hydroxy-6-oxooxan-2-yl]ethyl}-3,7-dimethyl-1,2,3,7,8,8a-hexahydronaphthalen-1-yl (2S)-2-methylbutanoate
|
|
| Clinical data | |
| Trade names | Mevacor |
| AHFS/Drugs.com | Monograph |
| MedlinePlus | a688006 |
| Pregnancy category |
|
| Routes of administration |
Oral |
| Legal status | |
| Legal status |
|
| Pharmacokinetic data | |
| Bioavailability | <5%[1] |
| Protein binding | >98%[1] |
| Metabolism | Hepatic (CYP3A andCYP2C8 substrate)[1] |
| Biological half-life | 2–5 hours[1] |
| Excretion | Faeces (83%), urine (10%)[1] |
| Identifiers | |
| CAS Number | 75330-75-5 |
| ATC code | C10AA02 (WHO) |
| PubChem | CID 53232 |
| IUPHAR/BPS | 2739 |
| DrugBank | DB00227 |
| ChemSpider | 48085 |
| UNII | 9LHU78OQFD |
| KEGG | D00359 |
| ChEBI | CHEBI:40303 |
| ChEMBL | CHEMBL503 |
| Synonyms | Monacolin K, Mevinolin |
| Chemical data | |
| Formula | C24H36O5 |
| Molar mass | 404.54 g/mol |
////////////////


 .
http://www.raps.org/Regulatory-Focus/News/2016/07/26/25394/FDA-Lowers-ANDA-Fee-Rates-for-2017/
Generic drugmakers submitting abbreviated new drug applications (ANDAs) and prior approval supplements (PAS) will see their US Food and Drug Administration (FDA) fee rates drop in 2017, though all other rates, including those for drug master files (DMF) and facility fees will increase when compared to 2016.
For FY 2017, the generic drug fee rates are: ANDA ($70,480, down from $76,030 in 2016), PAS ($35,240, down from $38,020 in 2016), DMF ($51,140, up from $42,170 in 2016), domestic active pharmaceutical ingredient (API) facility ($44,234, up from $40,867 in 2016), foreign API facility ($59,234, up from $55,867 in 2016), domestic finished dose formulation (FDF) facility ($258,646, up from $243,905), and foreign FDF facility ($273,646, up from $258,905 in 2016).
The new fees are effective 1 October 2016 and will remain in effect through 30 September 2017.
FDA explained the increases and decreases in fees, noting that for ANDA…
View original post 413 more words

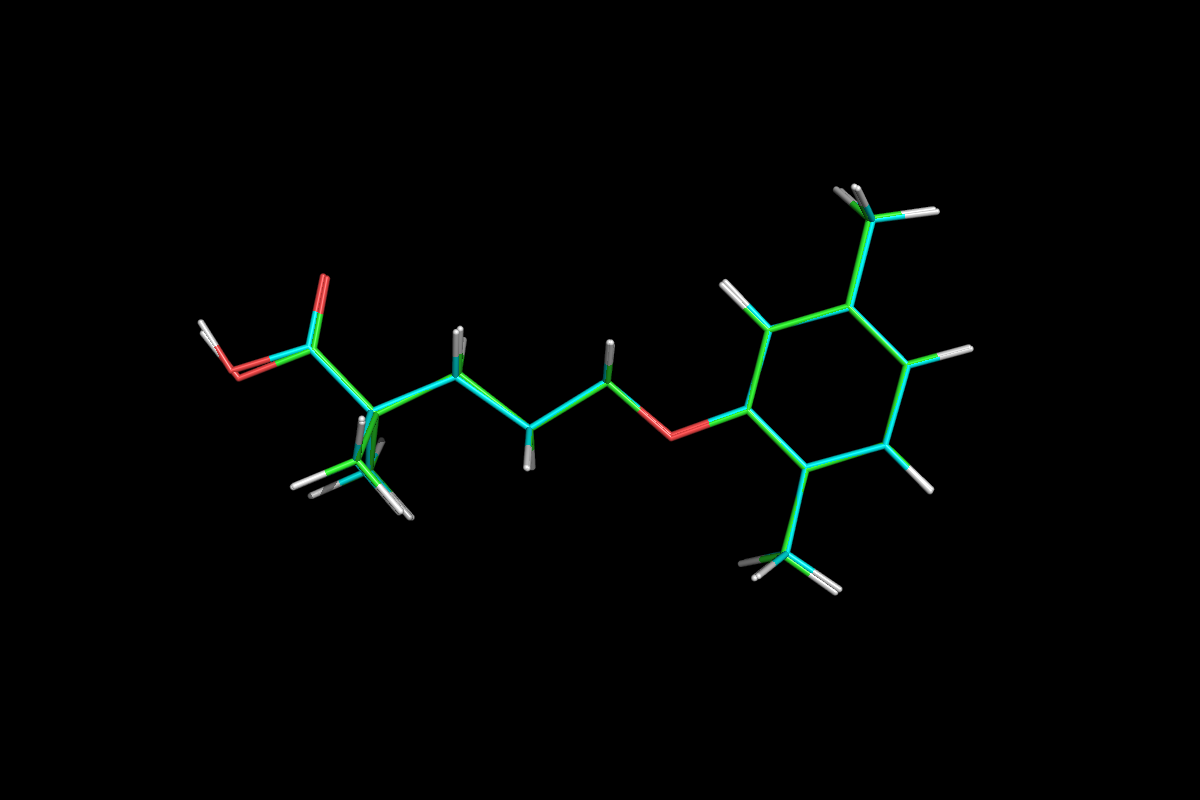
Gemfibrozil
5-(2,5-Dimethylphenoxy)-2,2-dimethylpentanoic Acid
IR (KBr, cm–1): 2959.03, 2919.78, 2877.65, 1709.42, 1613.44, 1586.60, 1511.07, 1473.81, 1414.01, 1387.89, 1317.61, 1286.34, 1271.91, 1214.39, 1159.26, 1048.83, 996.57, 803.75;
1H NMR (DMSO, 500 MHz, δ ppm): 1.12 (s, 6H), 1.60 and 1.67 (m, 4H), 2.08 (s, 3H), 2.24 (s, 3H), 3.90 (t, 2H), 6.62 (d, 1H), 6.70 (s, 1H), 6.97 (d, 1H);
13C NMR and DEPT (DMSO, 500 MHz, δ ppm): 15.39 (CH3), 20.94 (CH3), 24.67 (CH2), 24.87 (CH3, CH3), 36.43 (CH2), 40.91 (C), 67.57 (CH2), 112.07 (CH), 120.45 (CH), 122.44 (C), 129.96 (CH), 135.93 (C), 156.43 (C), 178.56 (C);
MS M/Z (ESI): 251.16 [(MH)+].
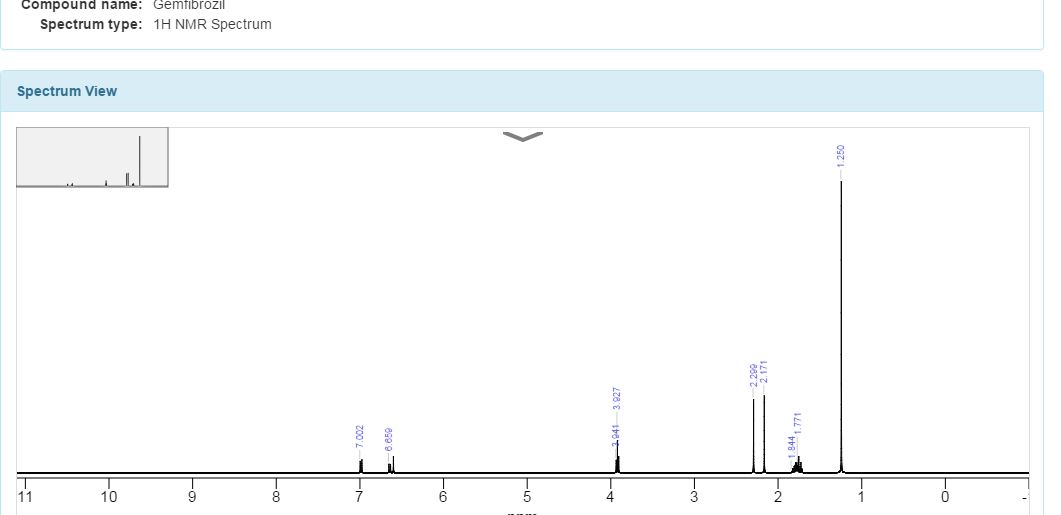
Solvent:CDCl3Instrument Type:JEOLNucleus:1HFrequency:400 MHzChemical Shift Reference:TMS


1H NMR spectrum of C15H22O3 in CDCL3 at 400 MHz
Gemfibrozil is the generic name for an oral drug used to lower lipid levels. It belongs to a group of drugs known as fibrates. It is most commonly sold as the brand name, Lopid. Other brand names include Jezil and Gen-Fibro.
Gemfibrozil was selected from a series of related compounds synthesized in the laboratories of the American company Parke Davisin the late 1970s. It came from research for compounds that lower plasma lipid levels in humans and in animals.[1]
Gemfibrozil has been detected in biosolids (the solids remaining after wastewater treatment) at concentrations up to 2650 ng/g wet weight.[3] This indicates that it survives the wastewater treatment process.
SYNTHESIS
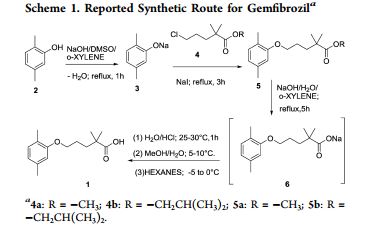


The sodium isobutyrate (I) is metallated with lithium diisopropylamide, and the resulting compound is alkylated with 3- (2,5-dimethylphenoxy) propyl bromide.
PATENT
Paul, L. C. 2,2-Dimethyl-ω-aryloxy alkanoic acids and salts and ester thereof. U.S. 3,674,836, 1972.
http://www.google.co.in/patents/US3674836
CLIP
Production of Gemfibrozil
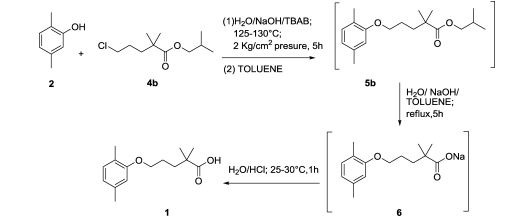
(1)2,5-Dimethylphenol and 1-Bromo-3-chloropropane reaction of 1-(2,5-dimethylphenoxy)-3-chloropropane. The reaction is carried out in toluene, adding new clean off reflux 5h. Just as follows:

(2)N/A can be used to manufacture Gemfibrozil.

PAPER
An improved process for the preparation of gemfibrozil, an antihypolipodimic drug substance, with an overall yield of 80% and ∼99.9% purity (including three chemical reactions) is reported. Formation and control of possible impurities are also described. Finally, gemfibrozil is isolated from water without any additional solvent purification.
Literature References:
Serum lipid regulating agent. Prepn: P. L. Creger, DE 1925423; eidem, US 3674836 (1969, 1972, both to Parke, Davis).
Production: O. P. Goel, US 4126637 (1978 to Warner-Lambert).
Pharmacology: A. H. Kissebach et al.,Atherosclerosis 24, 199 (1976); M. T. Kahonen et al., ibid. 32, 47 (1979).
Series of articles on metabolism, clinical pharmacology, kinetics and toxicology: Proc. R. Soc. Med. 69, Suppl 2, 1-120 (1976).
Toxicity data: S. M. Kurtz et al., ibid. 15.
Clinical trial in hyperlipidemia: J. E. Lewis et al., Pract. Cardiol. 9, 99 (1983).
Clinical reduction of cardiovascular risk in patients with low HDL levels: H. B. Rubins et al., N. Engl. J. Med. 341, 410 (1999).
. PMID 828263. |
|
| Systematic (IUPAC) name | |
|---|---|
|
5-(2,5-dimethylphenoxy)-2,2-dimethyl-pentanoic acid
|
|
| Clinical data | |
| Trade names | Lopid |
| AHFS/Drugs.com | Monograph |
| MedlinePlus | a686002 |
| Pregnancy category |
|
| Routes of administration |
Oral |
| Legal status | |
| Legal status |
|
| Pharmacokinetic data | |
| Bioavailability | Close to 100% |
| Protein binding | 95% |
| Metabolism | Hepatic (CYP3A4) |
| Biological half-life | 1.5 hours |
| Excretion | Renal 94% Feces 6% |
| Identifiers | |
| CAS Number | 25812-30-0 |
| ATC code | C10AB04 (WHO) |
| PubChem | CID 3463 |
| IUPHAR/BPS | 3439 |
| DrugBank | DB01241 |
| ChemSpider | 3345 |
| UNII | Q8X02027X3 |
| KEGG | D00334 |
| ChEBI | CHEBI:5296 |
| ChEMBL | CHEMBL457 |
| Chemical data | |
| Formula | C15H22O3 |
| Molar mass | 250.333 g/mol |
LOPID® (gemfibrozil tablets, USP) is a lipid regulating agent. It is available as tablets for oral administration. Each tablet contains 600 mg gemfibrozil. Each tablet also contains calcium stearate, NF; candelilla wax, FCC; microcrystalline cellulose, NF; hydroxypropyl cellulose, NF; hypromellose, USP; methylparaben, NF; Opaspray white; polyethylene glycol, NF; polysorbate 80, NF; propylparaben, NF; colloidal silicon dioxide, NF; pregelatinized starch, NF. The chemical name is 5-(2,5-dimethylphenoxy)2,2-dimethylpentanoic acid, with the following structural formula:
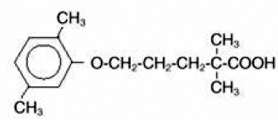 |
The empirical formula is C15H22O3 and the molecular weight is 250.35; the solubility in water and acid is 0.0019% and in dilute base it is greater than 1%. The melting point is 58° –61°C. Gemfibrozil is a white solid which is stable under ordinary conditions.
/////////Gemfibrozil, Antilipemic, Fibrates, 25812-30-0,
CC1=CC(OCCCC(C)(C)C(O)=O)=C(C)C=C1
GSK-2041706A
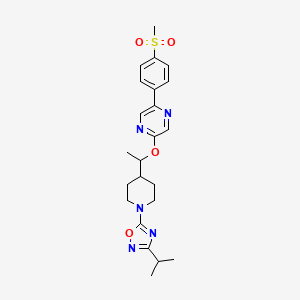
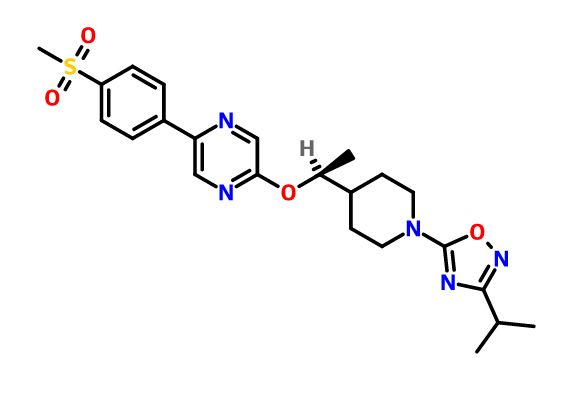
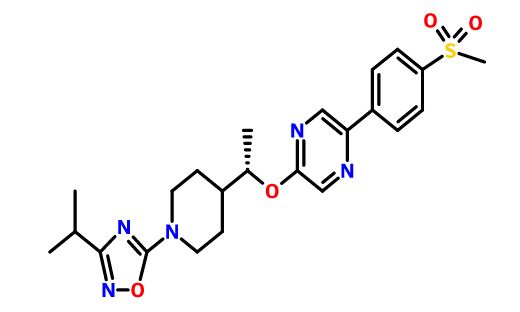
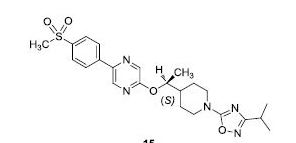
[2-([(1S)-1-(1-[3-(1-methylethyl)-1,2,4-oxadiazol-5-yl]-4-piperidinyl)ethyl]oxy)-5-[4-(methylsulfonyl)phenyl]pyrazine]
2-[((1S)-1-{1-[3-(1-Methylethyl)-1,2,4-oxadiazol-5-yl]-4-piperidinyl}ethyl)oxy]-5-[4-(methylsulfonyl)phenyl]pyrazine
Potent GPR119 Receptor Agonists
CAS 1032824-43-3
| Molecular Formula: | C23H29N5O4S |
|---|---|
| Molecular Weight: | 471.57246 g/mol |
G protein-coupled receptor 119 (GPR119) is a G protein-coupled receptor expressed predominantly in pancreatic β-cells and gastrointestinal enteroendocrine cells. Metformin is a first-line treatment of type 2 diabetes, with minimal weight loss in humans. In this study, we investigated the effects of GSK2041706 [2-([(1S)-1-(1-[3-(1-methylethyl)-1,2,4-oxadiazol-5-yl]-4-piperidinyl)ethyl]oxy)-5-[4-(methylsulfonyl)phenyl]pyrazine], a GPR119 agonist, and metformin as monotherapy or in combination on body weight in a diet-induced obese (DIO) mouse model. Relative to vehicle controls, 14-day treatment with GSK2041706 (30 mg/kg b.i.d.) or metformin at 30 and 100 mg/kg b.i.d. alone caused a 7.4%, 3.5%, and 4.4% (all P < 0.05) weight loss, respectively. The combination of GSK2041706 with metformin at 30 or 100 mg/kg resulted in a 9.5% and 16.7% weight loss, respectively. The combination of GSK2041706 and metformin at 100 mg/kg caused a significantly greater weight loss than the projected additive weight loss of 11.8%. This body weight effect was predominantly due to a loss of fat. Cumulative food intake was reduced by 17.1% with GSK2041706 alone and 6.6% and 8.7% with metformin at 30 and 100 mg/kg, respectively. The combination of GSK2041706 with metformin caused greater reductions in cumulative food intake (22.2% at 30 mg/kg and 37.5% at 100 mg/kg) and higher fed plasma glucagon-like peptide 1 and peptide tyrosine tyrosine levels and decreased plasma insulin and glucose-dependent insulinotropic polypeptide levels compared with their monotherapy groups. In addition, we characterized the effect of GSK2041706 and metformin as monotherapy or in combination on neuronal activation in the appetite regulating centers in fasted DIO mice. In conclusion, our data demonstrate the beneficial effects of combining a GPR119 agonist with metformin in the regulation of body weight in DIO mice.
Diabetes mellitus is an ever-increasing threat to human health. For example, in the United States current estimates maintain that about 16 million people suffer from diabetes mellitus.
Type I diabetes, also known as insulin-dependent diabetes mellitus (IDDM), is caused by the autoimmune destruction of the insulin producing pancreatic β-cells, and necessitates regular administration of exogenous insulin. Without insulin, cells cannot absorb sugar (glucose), which they need to produce energy. Symptoms of Type I diabetes usually start in childhood or young adulthood. People often seek medical help because they are seriously ill from sudden symptoms of high blood sugar (hyperglycemia).
Type II diabetes, also known as non-insulin-dependent diabetes mellitus (NIDDM), manifests with an inability to adequately regulate blood-glucose levels. Type II diabetes may be characterized by a defect in insulin secretion or by insulin resistance, namely those that suffer from Type II diabetes have too little insulin or cannot use insulin effectively. Insulin resistance refers to the inability of body tissues to respond properly to endogenous insulin. Insulin resistance develops because of multiple factors, including genetics, obesity, increasing age, and having high blood sugar over long periods of time. Type II diabetes, sometimes called mature or adult onset diabetes, can develop at any age, but most commonly becomes apparent during adulthood. The incidence of Type II diabetes in children, however, is rising
In diabetics, glucose levels build up in the blood and urine causing excessive urination, thirst, hunger, and problems with fat and protein metabolism. If left untreated, diabetes mellitus may cause life-threatening complications, including blindness, kidney failure, and heart disease.
Type II diabetes accounts for approximately 90-95% of diabetes cases, killing about 193,000 U.S. residents each year. Type II diabetes is the seventh leading cause of all deaths. In Western societies, Type II diabetes currently affects 6% of the adult population with world-wide frequency expected to grow by 6% per annum.
Although there are certain inheritable traits that may predispose particular individuals to developing Type II diabetes, the driving force behind the current increase in incidence of the disease is the increased sedentary lifestyle, diet, and obesity now prevalent in developed countries. About 80% of diabetics with Type II diabetes are significantly overweight. As noted above, an increasing number of young people are developing the disease. Type II diabetes is now internationally recognized as one of the major threats to human health in the 21stcentury.
Type II diabetes currently is treated at several levels. A first level of therapy is through the use of diet and/or exercise, either alone or in combination with therapeutic agents. Such agents may include insulin or pharmaceuticals that lower blood glucose levels. About 49% of individuals with Type II diabetes require oral medication(s), about 40% of individuals require insulin injections or a combination of insulin injections and oral medication(s), and about 10% of individuals may use diet and exercise alone.
Current therapies for diabetes mellitus include: insulin; insulin secretagogues, such as sulphonylureas, which increase insulin production from pancreatic-cells; glucose-lowering effectors, such as metformin which reduce glucose production from the liver; activators of the peroxisome proliferator-activated receptor—(PPAR-), such as the thiazolidinediones, which enhances insulin action; and α-glucosidase inhibitors which interfere with gut glucose production. There are, however, deficiencies associated with currently available treatments, including hypoglycemic episodes, weight gain, loss in responsiveness to therapy over time, gastrointestinal problems, and edema.
There are several areas at which research is being targeted in order to bring new, more effective, therapies to the marketplace. For example, on-going research includes exploring a reduction in excessive hepatic glucose production, enhancing the pathway by which insulin transmits its signal to the cells such that they take up glucose, enhancing glucose-stimulated insulin secretion from the pancreatic-cells, and targeting obesity and associated problems with fat metabolism and accumulation.
One particular target is GPR119. GPR119 is a member of the rhodopsin family of G-protein-coupled receptors. In addition to the “GPR119” identifier, several other identifiers exist, including but not limited to RUP 3, Snorf 25, 19 AJ, GPR 116 (believed to be erroneous), AXOR 20, and PS1. GPR119 is expressed in human gastrointestinal regions and in human islets. Activation of GPR119 has been demonstrated to stimulate intracellular cAMP and lead to glucose-dependent GLP-1 and insulin secretion. See, T. Soga et al., Biochemical and Biophysical Research Communications 326 (2005) 744-751, herein incorporated by reference with regard to a background understanding of GPR119.
In type 2 diabetes the action of GLP-1 on the β-cell is maintained, although GLP-1 secretion, itself, is reduced. More recently, therefore, much research has been focused on GLP-1. Studies show glucose-lowering effects in addition to GLP-1’s ability to stimulate glucose-dependent insulin secretion including, but not limited to, an inhibition of the release of the hormone glucagon following meals, a reduction in the rate at which nutrients are absorbed into the bloodstream, and a reduction of food intake. Studies demonstrate that treatments to increase GLP-1, therefore, may be used for a variety of conditions and disorders including but not limited to metabolic disorders, gastrointestinal disorders, inflammatory diseases, psychosomatic, depressive, and neuropsychiatric disease including but not limited to diabetes mellitus (Type 1 and Type 2), metabolic syndrome, obesity, appetite control and satiety, weight loss, stress, inflammation, myocardial ischemia/reperfusion injury, Alzheimer’s Disease, and other diseases of the central nervous system.
The use of exogenous GLP-1 in clinical treatment is severely limited, however, due to its rapid degradation by the protease DPP-IV. There are multiple GLP-1 mimetics in development for type 2 diabetes that are reported in the literature, all are modified peptides, which display longer half-lives than endogenous GLP-1. For example, the product sold under the tradename BYETTA® is the first FDA-approved agent of this new class of medications. These mimetics, however, require injection. An oral medication that is able to elevate GLP-1 secretion is desirable. Orally available inhibitors of DPP-IV, which result in elevation in intact GLP-1, are now available, such as sitagliptin, marketed under the brand name JANUVIA®. Nevertheless, a molecule which may stimulate GLP-1 secretion would provide a therapeutic benefit. A molecule which could stimulate both GLP-1 secretion and insulin secretion through effects on the L-cell and direct effects on the β-cell would hold much promise for type 2 diabetes therapy.
The present invention identifies agonists of GPR119 which increase glucose-disposal in part through elevation of GIP, GLP-1, and insulin. Moreover, studies demonstrate that GPR119 agonists such as the compounds of the present invention can stimulate incretins independently of glucose. GIP and GLP-1 are peptides, known as incretins, secreted from enteroendocrine K and L cells, respectively, in response to ingestion of nutrients, and have a wide variety of physiological effects that have been described in numerous publications over the past two decades. See, for example, Bojanowska, E. et al.,Med. Sci. Monit., 2005, August 11(8): RA271-8; Perry, T. et al., Curr. Alzheimer Res., 2005, July 2(3): 377-85; and Meier, J. J. et al.,Diabetes Metab. Res. Rev., 2005, March-April; 21(2); 91-117 (each herein incorporated by reference with regard to a background understanding of incretins). Moreover, although the mechanisms regulating GLP-1 secretion remain unclear, the initial rapid rise in GLP-1 following a meal may be a result of hormonal stimulation of neuronal afferents involving GIP. See, for example, J. N. Roberge and P. L. Brubaker, Endocrinology 133 (1993), pp. 233-240 (herein incorporated by reference with regard to such teaching). Furthermore, later increases in GLP-1 may involve direct activation of L-cells by nutrients in the distal small-intestine and the colon. GIP and GLP-1 are potent stimulators of the body’s ability to produce insulin in response to elevated levels of blood sugar. In Type 2 diabetes, patients display a decreased responsiveness to GIP but not GLP-1, with respect to its ability to stimulate insulin secretion. The mechanism behind the decreased responsiveness to GIP remains unclear since type 2 diabetics retain sensitivity to a bolus administration of GIP but not to a continuous infusion (Meier et al. 2004 Diabetes 53 S220-S224). Moreover recent studies with a long-acting fatty-acid derivative of GIP showed beneficial effects on glucose homeostasis in ob/ob mice following 14 days of treatment (Irwin N. et al. (2006) J. Med. Chem. 49, 1047-1054.)
Agonists to GPR119 may be of therapeutic value for diabetes and associated conditions, particularly type II diabetes, obesity, glucose intolerance, insulin resistance, metabolic syndrome X, hyperlipidemia, hypercholesterolemia, and atherosclerosis.
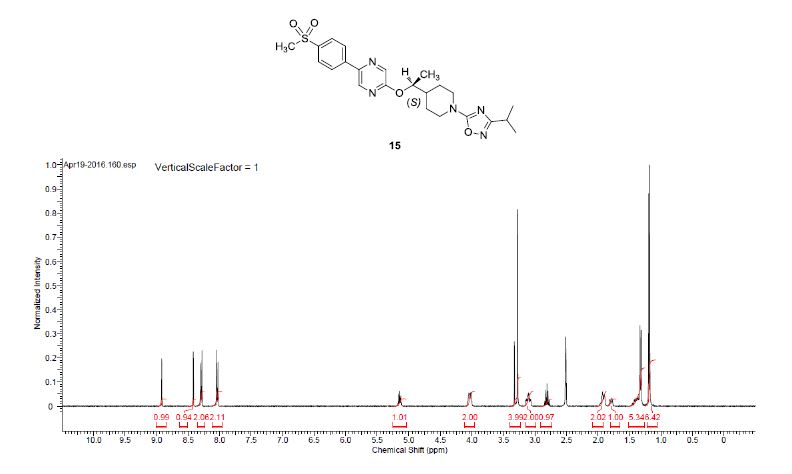
NMR
1H NMR (400 MHz, DMSO-d6) δ 8.91 (bs, 1H), 8.40 (bs, 1 H), 8.28 (d, J = 8.5 Hz, 2H), 8.02 (d, J = 8.5 Hz, 2H), 5.17–5.09 (m, 1H), 4.09–3.95 (m, 2H), 3.27 (s, 3H), 3.16–2.99 (m, 2H), 2.80 (q, J = 6.9 Hz, 1H), 1.98–1.85 (m, 2H), 1.83–1.70 (m, 1H), 1.47–1.33 (m, 2H), 1.31 (d, J = 6.3 Hz, 3H), 1.17 (d, J = 6.8 Hz, 6H).
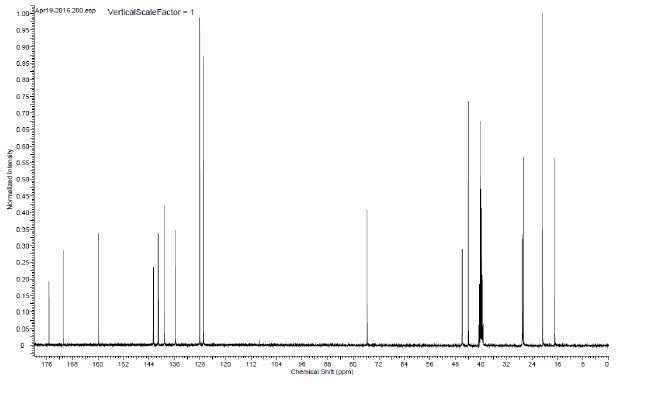
13C NMR (100.6 MHz, DMSO-d6) 175.3, 170.9, 159.8, 142.6, 141.2, 141.0, 139.1, 135.7, 128.1, 126.9, 75.7, 46.0, 45.9, 44.0, 40.2, 27.1, 27.0, 26.7, 20.7, 16.9.
HRMS calcd for C23H30N5O4S (M + H)+ 472.2013, found, 472.2009.
PATENT
Jing Fang, Jun Tang, Andrew J. Carpenter,Gregory Peckham, Christopher R. Conlee,Kien S. Du, Subba Reddy Katamreddy,
http://www.google.co.ug/patents/US20120077812
Example 156(±)-2-[(1-{1-[3-(1-Methylethyl)-1,2,4-oxadiazol-5-yl]-4-piperidinyl}ethyl)oxy]-5-[4-(methylsulfonyl)phenyl]pyrazine
Step 1: A solution of 3-(1-methylethyl)-5-(trichloromethyl)-1,2,4-oxadiazole (prepared as in Example 158, Alternative synthesis, Step 3, 179 g, 0.78 mol) in MeOH (300 mL) was treated with 4-piperidinemethanol (108 g, 0.94 mol) and stirred and heated at 50° C. overnight. The solvent was removed and the residue was purified by flash chromatography on a silica gel column to give {1-[3-(1-methylethyl)-1,2,4-oxadiazol-5-yl]-4-piperidinyl}methanol (60 g, 34%) as a pale yellow oil.
Step 2: A solution of {1-[3-(1-methylethyl)-1,2,4-oxadiazol-5-yl]-4-piperidinyl}methanol (1.50 g, 6.66 mmol) in CH2Cl2 (50 mL) at 0° C. was treated with Dess-Martin periodinane (2.91 g, 6.66 mmol). The reaction mixture was warmed to ambient temperature and stirred overnight. The reaction was quenched with aqueous 20% Na2S2O3(100 mL) and aqueous saturated NaHCO3 (100 mL) and then stirred for 10 minutes. The CH2Cl2 layer was separated and washed with brine, dried over Na2SO4, filtered, and the filtrate was concentrated to give the crude product as a cloudy colorless oil. The crude product was dissolved in 100 mL of 1:1 EtOAc/hexanes, filtered through a pad of silica gel, washed with 200 mL of 1:1 EtOAc/hexanes. The filtrate was concentrated to give 1.07 g (72%) of 1-[3-(1-methylethyl)-1,2,4-oxadiazol-5-yl]-4-piperidinecarbaldehyde as a clear colorless oil, which was used without further purification. 1H NMR (400 MHz, CDCl3): δ 9.68 (s, 1H), 4.15-4.00 (m, 2H), 3.30-3.20 (m, 2H), 2.86 (septet, 1H, J=7.0 Hz), 2.55-2.45 (m, 1H), 2.10-1.95 (m, 2H), 1.80-1.65 (m, 2H), 1.26 (d, 6H, J=6.8 Hz).
Step 3: (±)-1-{1-[3-(1-Methylethyl)-1,2,4-oxadiazol-5-yl]-4-piperidinyl}ethyl methanesulfonate (0.74 g, 49%) was prepared as a light brown oil from 1-[3-(1-methylethyl)-1,2,4-oxadiazol-5-yl]-4-piperidinecarbaldehyde (1.07 g, 4.79 mmol) and methylmagnesium bromide (3M in Et2O, 3.51 mL, 10.54 mmol) then methanesulfonyl chloride (0.22 mL, 2.81 mmol) and Et3N (0.66 mL, 4.68 mmol) in a manner similar to Example 139, Steps 1-2. The crude product was used without further purification. 1H NMR (400 MHz, CDCl3): δ 4.70-4.60 (m, 1H), 4.30-4.15 (m, 2H), 3.10-2.95 (m, 5H), 2.87 (septet, 1H, J=7.0 Hz), 1.95-1.70 (m, 3H), 1.55-1.35 (m, 5H), 1.26 (d, 6H, J=6.8 Hz).
Step 4: The title compound (0.212 g, 26%) was prepared as a white foam from 5-[4-(methylsulfonyl)phenyl]-2-pyrazinol (and tautomers thereof) (prepared as in Example 145, Steps 1-2, 0.43 g, 1.72 mmol), (±)-1-{1-[3-(1-methylethyl)-1,2,4-oxadiazol-5-yl]-4-piperidinyl}ethyl methanesulfonate (0.74 g, 2.32 mmol) and K2CO3 (0.48 g, 3.44 mmol) in DMF (15 mL) in a manner similar to Example 152, Steps 3. The crude product was purified by chromatography on an ISCO silica gel column using 0 to 25% EtOAc/CH2Cl2, followed by chromatography on a silica gel column eluted with 50% EtOAc/hexanes to give (±)-2-[(1-{1-[3-(1-methylethyl)-1,2,4-oxadiazol-5-yl]-4-piperidinyl}ethypoxy]-5-[4-(methylsulfonyl)phenyl]pyrazine as a white solid. 1H NMR (400 MHz, CDCl3): δ 8.53 (s, 1H), 8.25 (s, 1H), 8.10 (d, 2H, J=8.5 Hz), 8.02 (d, 2H, J=8.5 Hz), 5.20-5.10 (m, 1H), 4.35-4.20 (m, 2H), 3.15-3.00 (m, 5H), 2.91 (septet, 1H, J=7.0 Hz), 2.00-1.80 (m, 3H), 1.60-1.40 (m, 2H), 1.34 (d, 3H, J=6.1 Hz), 1.28 (d, 6H, J=7.1 Hz); LRMS (ESI), m/z 472 (M+H).
Example 1572-[((1R)-1-{1-[3-(1-Methylethyl)-1,2,4-oxadiazol-5-yl]-4-piperidinyl}ethyl)oxy]-5-[4-(methylsulfonyl)phenyl]pyrazin
The racemic 2-[(1-{1-[3-(1-methylethyl)-1,2,4-oxadiazol-5-yl]-4-piperidinyl}ethyl)oxy]-5-[4-(methylsulfonyl)phenyl]pyrazine (prepared as in Example 156) was subjected to Chiral HPLC [column: AS-H, column mobile phase: 70% CO2: 30% MeOH (2 mL/min), pressure 140 bar, temperature 40° C., 215 nm] analysis and then separated to give two (R and S) enantiomers. The title compound was isolated as an off-white solid with Tr of 23.42 min (first eluting peak). The (R) absolute stereochemistry was assigned by Ab initio VCD analysis.
Example 158
2-[((1S)-1-{1-[3-(1-Methylethyl)-1,2,4-oxadiazol-5-yl]-4-piperidinyl}ethyl)oxy]-5-[4-(methylsulfonyl)phenyl]pyrazine
The racemic 2-[(1-{1-[3-(1-methylethyl)-1,2,4-oxadiazol-5-yl]-4-piperidinyl}ethyl)oxy]-5-[4-(methylsulfonyl)phenyl]pyrazine (prepared as in Example 156) was subjected to Chiral HPLC [column: AS-H, column mobile phase: 70% CO2: 30% MeOH (2 mL/min), pressure 140 bar, temperature 40° C., 215 nm] analysis and then separated to give two (R and S) enantiomers. The title compound was isolated as an off-white solid with Tr of 25.83 min (second eluting peak). The (S) absolute stereochemistry was assigned by Ab initio VCD analysis. Alternative preparation from enantiomerically enriched material:
Step 1: Triethylamine (315 mL, 2.26 mol) was added dropwise to formic acid (150 mL, 3.91 mol) with overhead stirring while maintaining the internal temperature below 60° C. with ice-bath cooling. Neat 4-acetylpyridine (100 mL, 0.904 mol) was then added rapidly while maintaining the temperature below 50° C. Following this addition, the reaction was allowed to cool to 28° C. and the chiral ruthenium catalyst [N-[(1R,2R)-2-(amino-N)-1,2-diphenylethyl]-2,4,6-trimethylbenzenesulfonamidato-N]chloro[(1,2,3,4,5,6-n)-1-methyl-4-(1-methylethyl)benzene]ruthenium (CAS#177552-91-9; for catalyst preparation, see: Uematsu, N.; Fujii, A.; Hashiguchi, S.; Ikariya, T.; Noyori, R.; J. Am. Chem. Soc. 1996, 118, 4916-4917) (3 g, 4.46 mmol) was added. The mixture was stirred under house vacuum for 4 h and then overnight under an atmosphere of nitrogen. The reaction mixture was added dropwise to a stirred solution of 10% Na2CO3 (4 L) and then extracted with EtOAc (3×1 L). The combined EtOAc layers were washed once with brine (1 L), treated with MgSO4 and Darco G-60 decolorizing charcoal and filtered through a 100 g plug of silica gel washing with 10% MeOH/EtOAc (1 L). The filtrate was concentrated to provide a dark oil that crystallized upon standing. The solid was dissolved in warm t-butyl methyl ether (250 mL) and the warm solution was filtered to remove a small amount of insoluble material. The filtrate was allowed to stir with cooling to room temperature and then to −15° C. The solids were collected by filtration, washing with cold t-butyl methyl ether and heptane, and then dried under high vacuum to yield (1R)-1-(4-pyridinyl)ethanol as a dark beige solid (62 g, 52.9% yield). This solid material was 96% ee based on chiral HPLC(HPLC conditions: AS-H column, 5% MeOH/CO2, 40° C., 140 bar, 2 mL/min). The filtrate was combined with the insoluble solid from the crystallization and concentrated in vacuo to yield additional (1R)-1-(4-pyridinyl)ethanol as a dark oil (37.5 g, 32% yield). This oily material was 78% ee based on chiral HPLC (see HPLC conditions above). 1H NMR (400 MHz, DMSO-d6): δ 8.47-8.43 (m, 2H), 7.32-7.28 (m, 2H), 5.37 (d, 1H, J=4.4 Hz), 4.72-4.64 (m, 1H), 1.44 (d, 3H, J=6.6 Hz).
Step 2: A solution of (1R)-1-(4-pyridinyl)ethanol (37 g, 0.3 mol, 78% ee) in MeOH (2 L) was charged with PtO2 (5 g) under nitrogen atmosphere followed by acetic acid (19 mL). The mixture was evacuated and purged with hydrogen several times and then stirred under an atmosphere of hydrogen for 2 d at room temperature. The mixture was filtered to remove catalyst and the filtrate was concentrated in vacuo and triturated with EtOAc to yield a cream-colored solid which was collected by filtration. The filter cake was dissolved in MeOH (500 mL) and 50% NaOH (15.8 g) was added. The resulting solution was stirred at 25° C. for 30 min and concentrated. The resulting solid was triturated with Et2O (700 mL) and stirred at 25° C. for 30 min, the solids were removed by filtration and the filtrate was dried over MgSO4 and filtered again. The final filtrate was concentrated to yield (1R)-1-(4-piperidinyl)ethanol (22 g, 57% yield) as a light beige solid. 1H NMR (400 MHz, CDCl3): δ 3.50 (quint, 1H, J=6.3 Hz), 3.13-3.01 (m, 2H), 2.61-2.47 (m, 2H), 1.88 (br, 2H), 1.84-1.73 (m, 1H), 1.63-1.52 (m, 1H), 1.41-1.27 (m, 1H), 1.23-1.05 (m, 2H), 1.13 (d, 3H, J=6.2 Hz).
Step 3: A stirred solution of N-hydroxy-2-methylpropanimidamide (16.33 g, 160 mmol) in pyridine (16.81 mL, 208 mmol) and dichloromethane (165 mL) at −15° C. was treated with trichloroacetyl chloride (19.63 mL, 176 mmol) over 40 min. The reaction was allowed to warm to ambient temperature and stirred for 42 h. Water (100 mL) was added and the reaction was stirred for 30 min. The dichloromethane was removed and the residue was diluted with water (50 mL) and extracted with ether (300 mL). The ether layer was washed with water, dried over MgSO4 and concentrated to afford 3-(1-methylethyl)-5-(trichloromethyl)-1,2,4-oxadiazole (28.0 g, 76% yield) as an orange liquid.1H NMR (400 MHz, CDCl3): δ 3.13 (septet, 1H, J=7.0 Hz), 1.36 (d, 6H, J=7.0 Hz).
Step 4: A solution of 3-(1-methylethyl)-5-(trichloromethyl)-1,2,4-oxadiazole (25.8 g, 112 mmol) and (1R)-1-(4-piperidinyl)ethanol (13.4 g, 104 mmol) in MeOH (15 mL) was stirred at ambient temperature under a stream of nitrogen for 7 days. The reaction was diluted with MeOH (40 mL), cooled in an ice bath and 1N NaOH (25 mL) was added. The mixture was allowed to warm to ambient temperature and stir for 1 h. The reaction was partitioned in EtOAc (300 mL)/1N NaOH (75 mL) and the layers were separated. The aqueous layer was saturated with NaCl and extracted with EtOAc (200 mL). The combined EtOAc layers were dried over MgSO4, concentrated and placed under high vacuum for 18 h to afford (1R)-1-{1-[3-(1-methylethyl)-1,2,4-oxadiazol-5-yl]-4-piperidinyl}ethanol (16.75 g, 68%) as an orange oil. 1H NMR (400 MHz, CDCl3): δ 4.14 (m, 2H), 3.57 (quint, 1H, J=6.3 Hz), 2.98 (m, 2H), 2.83 (septet, 1H, J=7.0 Hz), 1.90 (m, 1H), 1.86 (br, 1H), 1.67 (m, 1H), 1.45 (m, 1H), 1.33 (m, 2H), 1.23 (d, 6H, J=7.0 Hz), 1.16 (d, 3H, J=6.3 Hz); LRMS (ESI), m/z 240 (M+H).
Step 5: A solution of (1R)-1-{1-[3-(1-methylethyl)-1,2,4-oxadiazol-5-yl]-4-piperidinyl}ethanol (1.68 g, 7.0 mmol) in dichloromethane (100 mL) at 0° C. was treated with Et3N (1.98 mL, 14.0 mmol) followed by methanesulfonyl chloride (0.66 mL, 8.4 mmol). The mixture was stirred at 0° C. for 1 h, then at room temperature for 2 h. The mixture was diluted with dichloromethane (50 mL), washed with 1M NaH2PO4 (75 mL×2) and brine, and dried over Na2SO4 and concentrated to give (1R)-1-{1-[3-(1-methylethyl)-1,2,4-oxadiazol-5-yl]-4-piperidinyl}ethyl methanesulfonate (2.23 g, 7.0 mmol, 100% yield) as a brown oil, which was used without further purification.
Step 6: A mixture of 5-[4-(methylsulfonyl)phenyl]-2-pyrazinol (and tautomers thereof) (prepared as in Example 145, Step 2, 1.3 g, 5.19 mmol), (1R)-1-{1-[3-(1-methylethyl)-1,2,4-oxadiazol-5-yl]-4-piperidinyl}ethyl methanesulfonate (2.23 g, 7.0 mmol, 70% ee) and K2CO3 (1.45 g, 10.4 mmol) in DMF (35 mL) was stirred at 100° C. in a preheated oil bath overnight. The mixture was cooled to ambient temperature, treated with water, and the mixture was extracted with EtOAc (75 mL×2). The combined organic extracts were washed with water, brine and dried over Na2SO4, filtered, and the filtrate was concentrated to a brown oil, which was by chromatography on a silica gel column eluted with 50% EtOAc/hexanes followed by chromatography on an ISCO silica gel column using 0 to 60% EtOAc/hexanes to give 2-[((1S)-1-{1-[3-(1-methylethyl)-1,2,4-oxadiazol-5-yl]-4-piperidinyl}ethyl)oxy]-5-[4-(methylsulfonyl)phenyl]pyrazine (0.73 g, 70% ee, 30%) as a white solid. The solid was subjected to chiral separation (similar to conditions used above for Example 158) to yield 0.30 g of the title compound as a white solid. 1H NMR (400 MHz, CDCl3): δ 8.53 (d, 1H, J=1.3 Hz), 8.25 (d, 1H, J=1.3 Hz), 8.10 (d, 2H, J=8.3 Hz), 8.02 (d, 2H, J=8.5 Hz), 5.20-5.10 (m, 1H), 4.35-4.20 (m, 2H), 3.15-3.00 (m, 5H), 2.90 (septet, 1H, J=7.0 Hz), 2.00-1.80 (m, 3H), 1.60-1.40 (m, 2H), 1.34 (d, 3H, J=6.3 Hz), 1.28 (d, 6H, J=6.9 Hz); LRMS (ESI), m/z 472 (M+H).
Paper
Richard T. Matsuoka*†, Eric E. Boros#, Andrew D. Brown†, Kae M. Bullock†, Will L. Canoy‡, Andrew J. Carpenter#, Jeremy D. Cobb†, Shannon E. Condon†, Nicole M. Deschamps†, Vassil I. Elitzin†, Greg Erickson†,Jing M. Fang#, David H. Igo§, Biren K. Joshi‡, Istvan W. Kaldor#, Mark B. Mitchell†, Gregory E. Peckham#, Daniel W. Reynolds‡, Matthew C. Salmon†, Matthew J. Sharp†, Elie A. Tabet#, Jennifer F. Toczko†, Lianming Michael Wu‡, and Xiao-ming M. Zhou†

Practical and scalable syntheses were developed that were used to prepare multikilogram batches of GSK1292263A (1) and GSK2041706A (15), two potent G protein-coupled receptor 119 (GPR119) agonists. Both syntheses employed relatively cheap and readily available starting materials, and both took advantage of an SNAr synthetic strategy.
/////////////GSK2041706A, GSK 2041706A, GSK-2041706A, GSK2041706, GSK 2041706, GSK-2041706
O=S(c4ccc(c3cnc(OC(C2CCN(c1nc(C(C)C)no1)CC2)C)cn3)cc4)(C)=O

OR

Anidulafungin
V-Echinocandin
| CAS Number | 166663-25-8 |
|---|---|
N-[(3S,6S,9S,11R,15S,18S,20R,21R,24S,25S,26S)-6-[(1S,2R)-1,2-dihydroxy-2-(4-hydroxyphenyl)ethyl]-11,20,21,25-tetrahydroxy-3,15-bis[(1R)-1-hydroxyethyl]-26-methyl-2,5,8,14,17,23-hexaoxo-1,4,7,13,16,22-hexaazatricyclo[22.3.0.09,13]heptacosan-18-yl]- 4-{4-[4-(pentyloxy)phenyl]phenyl}benzamide
1H NMR (700 MHz, d6-DMSO) δ 0.91 (t, 3H), 1.12 (d, 3H), 1.36 (m, 2H), 1.41 (m, 2H), 1.74 (p, 2H), 1.88 and 1.97 (overlapped, 2H), 3.85 (overlapped, 1H), 4.01 (t, 2H), 4.35 (overlapped, 1H), 4.44 (m, 1H), 4.76 (m, 1H), 4.80 (m, 1H), 5.02 (m, 1H), 5.07 (d, 1H), 5.52 (d, 1H), 7.04 (d, 1H), 7.66 (d, 1H), 7.74 (d, 1H), 7.80 (d, 1H), 7.82 (d, 1H), 7.97 (d, 1H), 8.01 (d, 1H), 8.14 (broad s, 1H), 8.60 (d, 1H). IR (cm−1)
KBr νmax; 3450 (O−H), 2932 (C−H), 2871 (C−H), 1632 (C═O), 1517 (Ar), 1488 (Ar), 1248 (C−O), 821 (C−H out-of-plane bending Ar 2 adj H’s).
Anidulafungin (brand names: Eraxis (in U.S. and Russia), Ecalta (in Europe)) is a semisynthetic echinocandin used as anantifungal drug. Anidulafungin was originally manufactured and submitted for FDA approval by Vicuron Pharmaceuticals.[1] Pfizeracquired the drug upon its acquisition of Vicuron in the fall of 2005.[2] Pfizer gained approval by the Food and Drug Administration(FDA) on February 21, 2006;[3] it was previously known as LY303366. Preliminary evidence indicates it has a similar safety profile tocaspofungin. Anidulafungin has proven efficacy against esophageal candidiasis, but its main use will probably be in invasive Candidainfection;[4][5][6] it may also have application in treating invasive Aspergillus infection. It is a member of the class of antifungal drugs known as the echinocandins; its mechanism of action is by inhibition of (1→3)-β-D-glucan synthase, an enzyme important to the synthesis of the fungal cell wall.
Anidulafungin significantly differs from other antifungals in that it undergoes chemical degradation to inactive forms at body pH and temperature. Because it does not rely on enzymatic degradation or hepatic or renal excretion, the drug is safe to use in patients with any degree of hepatic or renal impairment.[7]
Distribution: 30–50 L. Protein binding: 84%.
Anidulafungin is not evidently metabolized by the liver. This specific drug undergoes slow chemical hydrolysis to an open-ring peptide which lacks antifungal activity. The half-life of the drug is 27 hours. Thirty percent is excreted in the feces (10% as unchanged drug). Less than 1% is excreted in the urine.[8][9][10]
Anidulafungin inhibits glucan synthase, an enzyme important in the formation of (1→3)-β-D-glucan, a major fungal cell wall component. Glucan synthase is not present in mammalian cells, so it is an attractive target for antifungal activity.[11]
Anidulafungin is manufactured via semisynthesis. The starting material is echinocandin B (a lipopeptide fermentation product ofAspergillus nidulans or the closely related species, A. rugulosus), which undergoes deacylation (cleavage of the linoleoyl side chain) by the action of a deacylase enzyme from the bacterium Actinoplanes utahensis;[12] in three subsequent synthetic steps, including a chemical reacylation, the antifungal drug anidulafungin[11][13] is synthesized.
Aspergillus nidulans. Anidulafungin is an echinocandin, a class of antifungal drugs that inhibits the synthesis of 1,3-β-D-glucan, an essential component of fungal cell walls.
ERAXIS (anidulafungin) is 1-[(4R,5R)-4,5-dihydroxy-N -[[4“-(pentyloxy)[1,1′:4′,1”-terphenyl]-4-yl]carbonyl]-L-ornithine]echinocandin B. Anidulafungin is a white to off-white powder that is practically insoluble in water and slightly soluble in ethanol. In addition to the active ingredient, anidulafungin, ERAXIS for Injection contains the following inactive ingredients:
50 mg/vial – fructose (50 mg), mannitol (250 mg), polysorbate 80 (125 mg), tartaric acid (5.6 mg), and sodium hydroxide and/or hydrochloric acid for pH adjustment.
100 mg/vial – fructose (100 mg), mannitol (500 mg), polysorbate 80 (250 mg), tartaric acid (11.2 mg), and sodium hydroxide and/or hydrochloric acid for pH adjustment.
The empirical formula of anidulafungin is C58H73N7O17 and the formula weight is 1140.3. The structural formula is
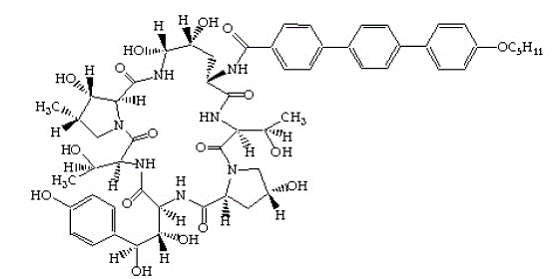
Prior to administration, ERAXIS for Injection requires reconstitution with sterile Water for Injection and subsequent dilution with either 5% DextroseInjection, USP or 0.9% Sodium Chloride Injection, USP (normal saline).
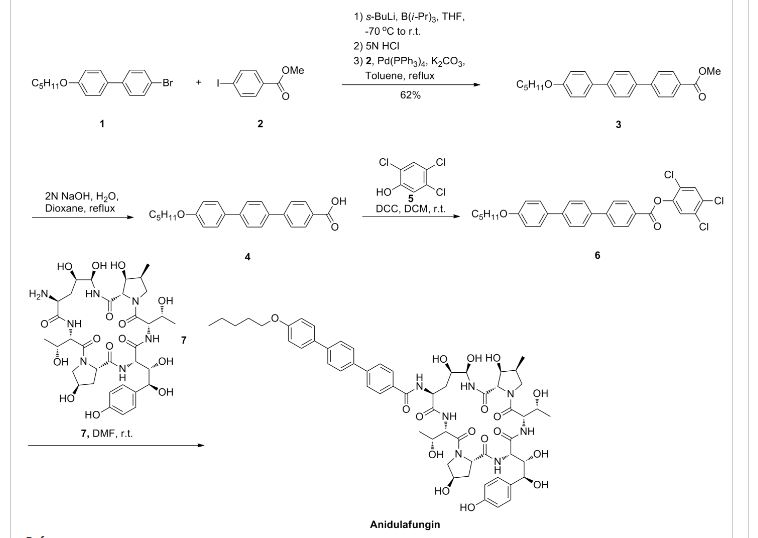
SYNTHESIS
J MED CHEM 1995, 38 3271-3281
pubs.acs.org/doi/abs/10.1021/jm00017a012
Aug 1, 1995 – J. Med. Chem. , 1995, 38 (17), pp 3271–3281. DOI: 10.1021/jm00017a012 … Journal ofMedicinal Chemistry 2001 44 (16), 2671-2674
Echinocandin B (ECB) is a lipopeptide composed of a complex cyclic peptide acylated at the N-terminus by linoleic acid. Enzymatic deacylation of ECB provided the peptide “nucleus” as a biologically inactive substrate from which novel ECB analogs were generated by chemical reacylation at the N-terminus. Varying the acyl group revealed that the structure and physical properties of the side chain, particularly its geometry and lipophilicity, played a pivotal role in determining the antifungal potency properties of the analog. Using CLOGP values to describe and compare the lipophilicities of the side chain fragments, it was shown that values of > 3.5 were required for expression of antifungal activity. Secondly, a linearly rigid geometry of the side chain was the most effective shape in enhancing the antifungal potency. Using these parameters as a guide, a variety of novel ECB analogs were synthesized which included arylacyl groups that incorporated biphenyl, terphenyl, tetraphenyl, and arylethynyl groups. Generally the glucan synthase inhibition by these analogs correlated well with in vitro and in vivo activities and was likewise influenced by the structure of the side chain. These structural variations resulted in enhancement of antifungal activity in both in vitro and in vivo assays. Some of these analogs, including LY303366 (14a), were effective by the oral route of administration.
PATENT
US 5965525
http://www.google.co.in/patents/US5965525
PATENT
US 4293482
http://www.google.co.in/patents/US4293482
Paper
http://pubs.acs.org/doi/abs/10.1021/op800055h

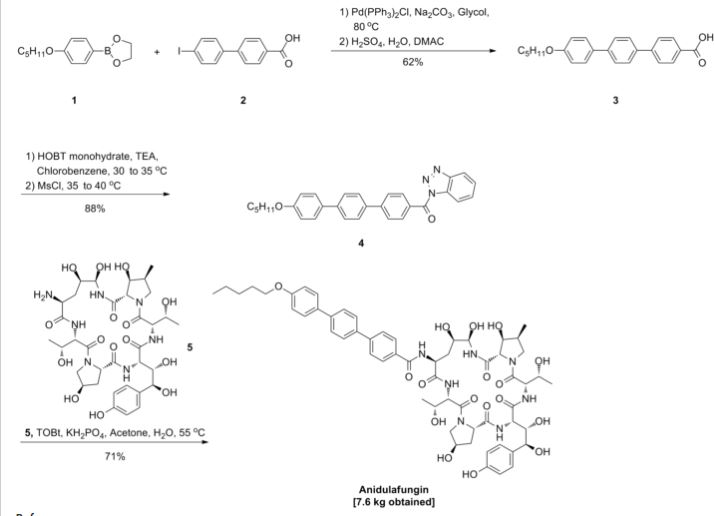
Many years ago anidulafungin 1 was identified as a potentially useful medicine for the treatment of fungal infections. Its chemical and physical properties as a relatively high molecular weight semisynthetic derived from echinocandin B proved to be a significant hurdle to its final presentation as a useful medicine. It has recently been approved as an intravenous treatment for invasive candidaisis, an increasingly common health hazard that is potentially life-threatening. The development and commercialization of this API, which is presented as a molecular mixture of anidulafungin and d-fructose is described. This includes, single crystal X-ray structures of the starting materials, the echinocandin B cyclic-peptide nucleus (ECBN·HCl) and the active ester 1-({[4′′-(pentyloxy)-1,1′:4′,1′′-terphenyl-4-yl]carbonyl}oxy)-1H-1,2,3-benzotriazole (TOBt). Details of the structure and properties of starting materials, scale-up chemistry and unusual crystallization phenomena associated with the API formation are discussed.
References
 |
|
| Systematic (IUPAC) name | |
|---|---|
|
N-[(3S,6S,9S,11R,15S,18S,20R,21R,24S,25S,26S)-6-[(1S,2R)-1,2-dihydroxy-2-(4-hydroxyphenyl)ethyl]-11,20,21,25-tetrahydroxy-3,15-bis[(1R)-1-hydroxyethyl]-26-methyl-2,5,8,14,17,23-hexaoxo-1,4,7,13,16,22-hexaazatricyclo[22.3.0.09,13]heptacosan-18-yl]- 4-{4-[4-(pentyloxy)phenyl]phenyl}benzamide
|
|
| Clinical data | |
| Trade names | Eraxis |
| AHFS/Drugs.com | Monograph |
| Pharmacokinetic data | |
| Protein binding | 84 % |
| Biological half-life | 40–50 hours |
| Identifiers | |
| CAS Number | 166663-25-8 |
| ATC code | J02AX06 (WHO) |
| PubChem | CID 166548 |
| DrugBank | DB00362 |
| ChemSpider | 21106258 |
| UNII | 9HLM53094I |
| KEGG | D03211  |
| ChEBI | CHEBI:55346 |
| ChEMBL | CHEMBL1630215 |
| Chemical data | |
| Formula | C58H73N7O17 |
| Molar mass | 1140.24 g/mol |
//////////FUNGIN, ANIDULAFUNGIN, Eraxis , Ecalta, semisynthetic echinocandin, anantifungal drug, FDA 2006, PFIZER, LY-307853, LY-329960, LY-333006, LY303366, VEC, VER-002, 166663-25-8, Eli Lilly and Company Inc.
CCCCCOc1ccc(cc1)c2ccc(cc2)c3ccc(cc3)C(=O)N[C@H]6C[C@@H](O)[C@@H](O)NC(=O)C4[C@@H](O)[C@@H](C)CN4C(=O)C(NC(=O)C(NC(=O)C5C[C@@H](O)CN5C(=O)C(NC6=O)[C@@H](C)O)[C@@H](O)[C@H](O)c7ccc(O)cc7)[C@@H](C)O
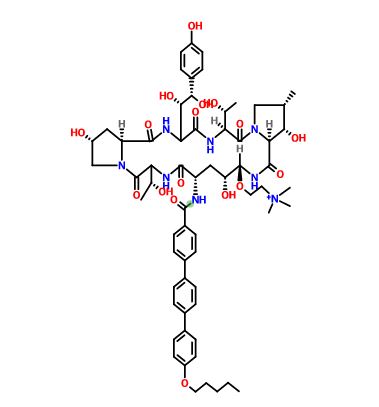
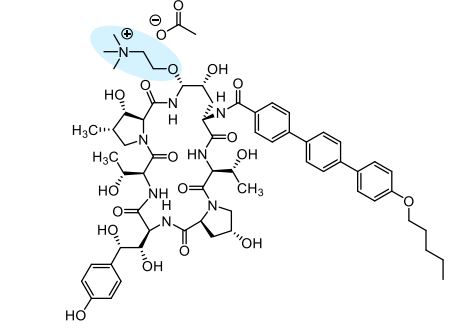
 as CH3COOH salt
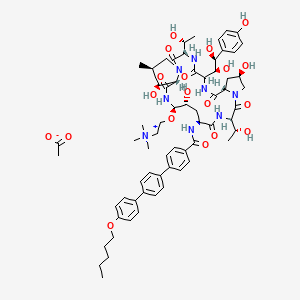
Several structural representations above
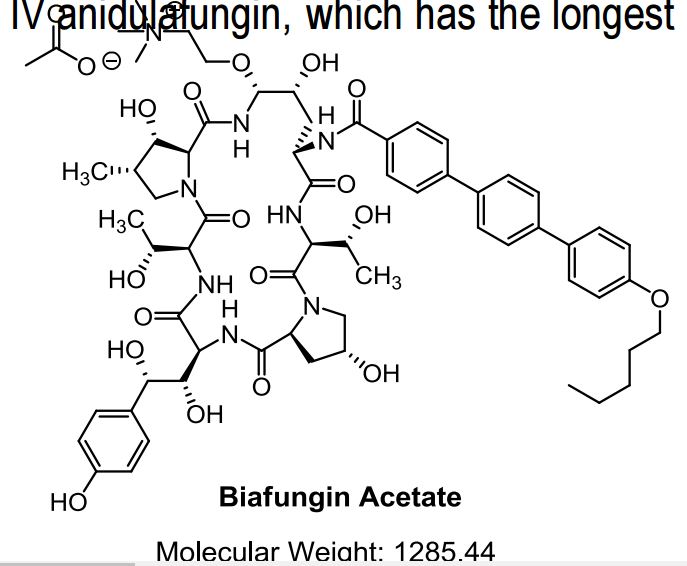
Biafungin™; CD 101 IV; CD 101 Topical; CD101; SP 3025, Biafungin acetate, Echinocandin B
UNII-G013B5478J FRE FORM,
CAS 1396640-59-7 FREE FORM
MF, C63-H85-N8-O17, MW, 1226.4035
| Treat and prevent invasive fungal infections; Treat and prevent systemic Candida infections; Treat candidemia |
Biafungin acetate
CAS 1631754-41-0 ACETATE, Molecular Formula, C63-H85-N8-O17.C2-H3-O2, Molecular Weight, 1285.4472,
UNII: W1U1TMN677
CD101 – A novel echinocandin antifungal C. albicans (n=351) MIC90 = 0.06 µg/mL C. glabrata (n=200) MIC90 = 0.06 µg/mL Echinocandins have potent fungicidal activity against Candida species
U.S. – Fast Track (Treat candidemia);
U.S. – Fast Track (Treat and prevent invasive fungal infections);
U.S. – Orphan Drug (Treat and prevent invasive fungal infections);
U.S. – Orphan Drug (Treat candidemia);
U.S. – Qualified Infectious Disease Program (Treat candidemia);
U.S. – Qualified Infectious Disease Program (Treat and prevent invasive fungal infections)
Fungal infections have emerged as major causes of human disease, especially among the immunocompromised patients and those hospitalized with serious underlying disease. As a consequence, the frequency of use of systemic antifungal agents has increased significantly and there is a growing concern about a shortage of effective antifungal agents. Although resistance rates to the clinically available antifungal agents remains low, reports of breakthrough infections and the increasing prevalence of uncommon fungal species that display elevated MIC values for existing agents is worrisome. Biafungin (CD101, previously SP 3025) is a novel echinocandin that displays chemical stability and long-acting pharmacokinetics that is being developed for once-weekly or other intermittent administration (see posters #A-693 and A- 694 for further information). In this study, we test biafungin and comparator agents against a collection of common Candida and Aspergillus species, including isolates resistant to azoles and echinocandins.
The echinocandins are an important class of antifungal agents, but are administered once daily by intravenous (IV) infusion. An echinocandin that could be administered once weekly could facilitate earlier hospital discharges and could expand usage to indications where daily infusions are impractical. Biafungin is a highly stable echinocandin for once-weekly IV administration. The compound was found to have a spectrum of activity and potency comparable to other echinocandins. In chimpanzees single dose pharmacokinetics of IV and orally administered biafungin were compared to IV anidulafungin, which has the longest half-life (T1/2 ) of the approved echinocandins.
Background Vulvovaginal candidiasis (VVC) is a highly prevalent mucosal infection VVC is caused by Candida albicans (~85%) and non-albicans (~15%) 5-8% of women have recurrent VVC (RVVC) which is associated with a negative impact on work/social life Oral fluconazole prescribed despite relapse, potential DDIs and increased risk to pregnant women No FDA-approved therapy for RVVC and no novel agent in >20 years


Cidara Therapeutics 6310 Nancy Ridge Drive, Suite 101 San Diego, CA 92121
The incidence of invasive fungal infections, especially those due to Aspergillus spp. and Candida spp., continues to increase. Despite advances in medical practice, the associated mortality from these infections continues to be substantial. The echinocandin antifungals provide clinicians with another treatment option for serious fungal infections. These agents possess a completely novel mechanism of action, are relatively well-tolerated, and have a low potential for serious drug–drug interactions. At the present time, the echinocandins are an option for the treatment of infections due Candida spp (such as esophageal candidiasis, invasive candidiasis, and candidemia). In addition, caspofungin is a viable option for the treatment of refractory aspergillosis. Although micafungin is not Food and Drug Administration-approved for this indication, recent data suggests that it may also be effective. Finally, caspofungin- or micafungin-containing combination therapy should be a consideration for the treatment of severe infections due to Aspergillus spp. Although the echinocandins share many common properties, data regarding their differences are emerging at a rapid pace. Anidulafungin exhibits a unique pharmacokinetic profile, and limited cases have shown a potential far activity in isolates with increased minimum inhibitory concentrations to caspofungin and micafungin. Caspofungin appears to have a slightly higher incidence of side effects and potential for drug–drug interactions. This, combined with some evidence of decreasing susceptibility among some strains ofCandida, may lessen its future utility. However, one must take these findings in the context of substantially more data and use with caspofungin compared with the other agents. Micafungin appears to be very similar to caspofungin, with very few obvious differences between the two agents.
Echinocandins are a new class of antifungal drugs[1] that inhibit the synthesis of glucan in the cell wall, via noncompetitive inhibition of the enzyme 1,3-β glucan synthase[2][3] and are thus called “penicillin of antifungals”[4] (a property shared with papulacandins) as penicillin has a similar mechanism against bacteria but not fungi. Beta glucans are carbohydrate polymers that are cross-linked with other fungal cell wall components (The bacterial equivalent is peptidoglycan). Caspofungin, micafungin, and anidulafungin are semisynthetic echinocandin derivatives with clinical use due to their solubility, antifungal spectrum, and pharmacokinetic properties.[5]


List of echinocandins:[17]
Discovery of echinocandins stemmed from studies on papulacandins isolated from a strain of Papularia sphaerosperma (Pers.), which were liposaccharide – i.e., fatty acid derivatives of a disaccharide that also blocked the same target, 1,3-β glucan synthase – and had action only on Candida spp. (narrow spectrum). Screening of natural products of fungal fermentation in the 1970s led to the discovery of echinocandins, a new group of antifungals with broad-range activity against Candida spp. One of the first echinocandins of the pneumocandin type, discovered in 1974, echinocandin B, could not be used clinically due to risk of high degree of hemolysis. Screening semisynthetic analogs of the echinocandins gave rise to cilofungin, the first echinofungin analog to enter clinical trials, in 1980, which, it is presumed, was later withdrawn for a toxicity due to the solvent system needed for systemic administration. The semisynthetic pneumocandin analogs of echinocandins were later found to have the same kind of antifungal activity, but low toxicity. The first approved of these newer echinocandins was caspofungin, and later micafungin and anidulafungin were also approved. All these preparations so far have low oral bioavailability, so must be given intravenously only. Echinocandins have now become one of the first-line treatments for Candida before the species are identified, and even as antifungal prophylaxis in hematopoietic stem cell transplant patients.
CIDARA THERAPEUTICS DOSES FIRST PATIENT IN PHASE 2 TRIAL OF CD101 TOPICAL TO TREAT VULVOVAGINAL CANDIDIASIS
SAN DIEGO–(BUSINESS WIRE)–Jun. 9, 2016– Cidara Therapeutics, Inc. (Nasdaq:CDTX), a biotechnology company developing novel anti-infectives and immunotherapies to treat fungal and other infections, today announced that the first patient has been dosed in RADIANT, a Phase 2 clinical trial comparing the safety and tolerability of the novel echinocandin, CD101, to standard-of-care fluconazole for the treatment of acute vulvovaginal candidiasis (VVC). RADIANT will evaluate two topical formulations of CD101, which is Cidara’s lead antifungal drug candidate.
“There have been no novel VVC therapies introduced for more than two decades, so advancing CD101 topical into Phase 2 is a critical step for women with VVC and for Cidara,” said Jeffrey Stein, Ph.D., president and chief executive officer of Cidara. “Because of their excellent safety record and potency against Candida, echinocandin antifungals are recommended as first line therapy to fight systemic Candida infections. CD101 topical will be the first echinocandin tested clinically in VVC and we expect to demonstrate safe and improved eradication of Candida with rapid symptom relief for women seeking a better option over the existing azole class of antifungals.”
RADIANT is a Phase 2, multicenter, randomized, open-label, active-controlled, dose-ranging trial designed to evaluate the safety and tolerability of CD101 in women with moderate to severe episodes of VVC. The study will enroll up to 125 patients who will be randomized into three treatment cohorts. The first cohort will involve the treatment of 50 patients with CD101 Ointment while a second cohort of 50 patients will receive CD101 Gel. The third cohort will include 25 patients who will be treated with oral fluconazole.
The primary endpoints of RADIANT will be the safety and tolerability of a single dose of CD101 Ointment and multiple doses of CD101 Gel in patients with acute VVC. Secondary endpoints include therapeutic efficacy in acute VVC patients treated with CD101. Treatment evaluations and assessments will occur on trial days 7, 14 and 28.
The RADIANT trial will be conducted at clinical trial centers across the United States. More information about the trial is available at www.clinicaltrials.gov, identifier NCT02733432.
About VVC and RVVC
Seventy-five percent of women worldwide suffer from VVC in their lifetime, and four to five million women in the United Statesalone have the recurrent form of the infection, which is caused by Candida. Many women will experience recurrence after the completion of treatment with existing therapies. Most VVC occurs in women of childbearing potential (the infection is common in pregnant women), but it affects women of all ages. In a recent safety communication, the U.S. Food and Drug Administration(FDA) advised caution in the prescribing of oral fluconazole for yeast infections during pregnancy based on a published study concluding there is an increased risk of miscarriage. The Centers for Disease Control and Prevention (CDC) guidelines recommend using only topical antifungal products to treat pregnant women with vulvovaginal yeast infections. Vaginal infections are associated with a substantial negative impact on day-to-day functioning and adverse pregnancy outcomes including preterm delivery, low birth weight, and increased infant mortality in addition to predisposition to HIV/AIDS. According to the CDC, certain species of Candida are becoming increasingly resistant to existing antifungal medications. This emerging resistance intensifies the need for new antifungal agents.
About CD101 Topical
CD101 topical is the first topical agent in the echinocandin class of antifungals and exhibits a broad spectrum of fungicidal activity against Candida species. In May 2016, the FDA granted Qualified Infectious Disease Product (QIDP) and Fast Track Designation to CD101 topical for the treatment of VVC and the prevention of RVVC.
About Cidara Therapeutics
Cidara is a clinical-stage biotechnology company focused on the discovery, development and commercialization of novel anti-infectives for the treatment of diseases that are inadequately addressed by current standard-of-care therapies. Cidara’s initial product portfolio comprises two formulations of the company’s novel echinocandin, CD101. CD101 IV is being developed as a once-weekly, high-exposure therapy for the treatment and prevention of serious, invasive fungal infections. CD101 topical is being developed for the treatment of vulvovaginal candidiasis (VVC) and the prevention of recurrent VVC (RVVC), a prevalent mucosal infection. In addition, Cidara has developed a proprietary immunotherapy platform, Cloudbreak™, designed to create compounds that direct a patient’s immune cells to attack and eliminate pathogens that cause infectious disease. Cidara is headquartered inSan Diego, California. For more information, please visit www.cidara.com.
REF http://ir.cidara.com/phoenix.zhtml?c=253962&p=irol-newsArticle&ID=2176474
CLIP
Cidara Therapeutics (formerly K2 Therapeutics) grabbed $42 million in a private Series B funding round Wednesday to continue developing its once-weekly anti-fungal therapy. Just in June 2014, the company completed a $32 million Series A financing led by 5AM Ventures, Aisling Capital, Frazier Healthcare and InterWest Partners, which was the fourth largest A round in 2014 for innovative startups[1]. FierceBiotech named the company as one of 2014 Fierce 15 biotech startups.
Cidara has an impressive executive team. The company was co-founded by Kevin Forrest, former CEO of Achaogen (NASDAQ: AKAO), and Shaw Warren. Jeffrey Stein, former CEO of Trius Therapeutics (NASDAQ: TSRX) and Dirk Thye, former president of Cerexa, have joined Cidara as CEO and CMO, respectively. Trius successfully developed antibiotic tedizolid and was acquired in 2013 by Cubist Pharmaceuticals (NASDAQ: CBST) for $818 million.
Cidara’s lead candidate, biafungin (SP3025), was acquired from Seachaid Pharmaceuticals for $6 million. Biafungin’s half-life is much longer than that of similar drugs known as echinocandins (e.g., caspofungin, micafungin, anidulafungin), which may allow it to be developed as a once-weekly therapy, instead of once daily. The company is also developing a topical formulation of biafungin, namely topifungin. Cidara intends to file an IND and initiate a Phase I clinical trial in the second half of 2015.
Merck’s Cancidas (caspofungin), launched in 2001, was the first of approved enchinocandins. The drug generated annual sales of $596 million in 2008. The approved echinocandins must be administered daily by intravenous infusion. Biafungin with improved pharmacokinetic characteristics has the potential to bring in hundreds of millions of dollars per year.
[1] Nat Biotechnol. 2015, 33(1), 18.
CLIP
Biafungin is a potent and broad-spectrum antifungal agent with excellent activity against wild-type and troublesome azole- and echinocandin-resistant strains of Candida spp. The activity of biafungin is comparable to anidulafungin. • Biafungin was active against both wild-type and itraconazole-resistant strains of Aspergillus spp. from four different species. • In vitro susceptibility testing of biafungin against isolates of Candida and Aspergillus may be accomplished by either CLSI or EUCAST broth microdilution methods each providing comparable results. • The use of long-acting intravenous antifungal agents that could safely be given once a week to select patients is desirable and might decrease costs with long-term hospitalizations. Background: A novel echinocandin, biafungin, displaying long-acting pharmacokinetics and chemical stability is being developed for once-weekly administration. The activities of biafungin and comparator agents were tested against 173 fungal isolates of the most clinically common species. Methods: 106 CAN and 67 ASP were tested using CLSI and EUCAST reference broth microdilution methods against biafungin (50% inhibition) and comparators. Isolates included 27 echinocandin-resistant CAN (4 species) with identified fks hotspot (HS) mutations and 20 azole nonsusceptible ASP (4 species). Results: Against C. albicans, C. glabrata and C. tropicalis, the activity of biafungin (MIC50, 0.06, 0.12 and 0.03 μg/ml, respectively by CLSI method) was comparable to anidulafungin (AND; MIC50, 0.03, 0.12 and 0.03 μg/ml, respectively) and caspofungin (CSP; MIC50, 0.12, 0.25 and 0.12 μg/ml, respectively; Table). C. krusei strains were very susceptible to biafungin, showing MIC90 values of 0.06 μg/ml by both methods. Biafungin (MIC50/90, 1/2 μg/ml) was comparable to AND and less potent than CSP against C. parapsilosis using CLSI methodology. CLSI and EUCAST methods displayed similar results for most species, but biafungin (MIC50, 0.06 μg/ml) was eight-fold more active than CSP (MIC50, 0.5 μg/ml) against C. glabrata using the EUCAST method. Overall, biafungin was two- to four-fold more active against fks HS mutants than CSP and results were comparable to AND. Biafungin was active against A. fumigatus (MEC50/90, ≤0.008/0.015 μg/ml), A. terreus (MEC50/90, 0.015/0.015 μg/ml), A. niger (MEC50/90, ≤0.008/0.03 μg/ml) and A. flavus (MEC50/90, ≤0.008/≤0.008 μg/ml) using CLSI method. EUCAST results for ASP were also low for all echinocandins and comparable to CLSI results. Conclusions: Biafungin displayed comparable in vitro activity with other echinocandins against common wild-type CAN and ASP and resistant subsets that in combination with the long-acting profile warrants further development of this compound. 1. Arendrup MC, Cuenca-Estrella M, Lass-Florl C, Hope WW (2013). Breakpoints for antifungal agents: An update from EUCAST focussing on echinocandins against Candida spp. and triazoles against Aspergillus spp. Drug Resist Updat 16: 81-95. 2. Castanheira M, Woosley LN, Messer SA, Diekema DJ, Jones RN, Pfaller MA (2014). Frequency of fks mutations among Candida glabrata isolates from a 10-year global collection of bloodstream infection isolates. Antimicrob Agents Chemother 58: 577-580. 3. Clinical and Laboratory Standards Institute (2008). M27-A3. Reference Method for Broth Dilution Antifungal Susceptibility Testing of Yeasts: third edition. Wayne, PA: CLSI. 4. Clinical and Laboratory Standards Institute (2008). M38-A2. Reference Method for Broth Dilution Antifungal Susceptibility Testing of Filamentous Fungi: Second Edition. Wayne, PA: CLSI. 5. Clinical and Laboratory Standards Institute (2012). M27-S4. Reference Method for Broth Dilution Antifungal Susceptibility Testing of Yeasts: 4th Informational Supplement. Wayne, PA: CLSI. 6. European Committee on Antimicrobial Susceptibility Testing (2014). Breakpoint tables for interpretation of MICs and zone diameters. Version 4.0, January 2014. Available at: http://www.eucast.org/clinical_breakpoints/. Accessed January 1, 2014. 7. Pfaller MA, Diekema DJ (2010). Epidemiology of invasive mycoses in North America. Crit Rev Microbiol 36: 1-53. 8. Pfaller MA, Diekema DJ, Andes D, Arendrup MC, Brown SD, Lockhart SR, Motyl M, Perlin DS (2011). Clinical breakpoints for the echinocandins and Candida revisited: Integration of molecular, clinical, and microbiological data to arrive at species-specific interpretive criteria. Drug Resist Updat 14: 164-176. ABSTRACT Activity of a Novel Echinocandin Biafungin (CD101) Tested against Most Common Candida and Aspergillus Species, Including Echinocandin- and Azole-resistant Strains M CASTANHEIRA, SA MESSER, PR RHOMBERG, RN JONES, MA PFALLER JMI Laboratories, North Liberty, Iowa, USA C
https://www.google.com/patents/WO2015035102A2?cl=en
BIAFUNGIN ACETATE IS USED AS STARTING MATERIAL
Example 30b: Synthesis of Compound 31


Step a. Nitration of Biafungin Acetate

To a stirring solution of biafungin (1 00 mg, 0.078 mmol) in glacial acetic acid(1 .5 ml_) was added sodium nitrite (1 1 mg, 0.159 mmol) and the reaction was stirred at ambient temperature for 20 hours. The mixture was applied directly to reversed phase H PLC (Isco CombiFlash Rf; 50g RediSep C1 8 column, 5 to 95% acetonitrile in Dl water containing 0.1 % formic acid: 15 minute gradient). The pure fractions were pooled and lyophilized to yield 85 mg of the desired product as a light yellow solid, formate salt. 1 H-NMR (300 M Hz, Methanol-d4) δ 8.58 (d, 1 H, J = 1 1 .7 Hz), 8.47 (t, 2H, J = 8.7Hz), 8.05 (d, 1 H, J = 2.1 Hz), 7.99 (d, 2H, J = 9.3 Hz), 7.82 (d, 2H, J = 8.7 Hz), 7.79-7.60 (m, 12H), 7.1 7 (d, 1 H, J = 8.7 Hz), 7.03 (d, 2H, J = 9 Hz), 5.48 (d, 1 H, J = 6 Hz), 5.08 (dd, 1 H, J = 1 .2, 5.7 Hz), 4.95-4.73 (m, 5H), 4.68-4.56 (m, 2H), 4.53 (d, 1 H, J = 5.7 Hz), 4.48-4.39 (m, 2H), 4.31 -3.79 (m, 6H), 4.04 (t, 2H, J = 5.7 Hz), 3.72-3.44 (m,3H), 3.1 8 (s, 9H), 2.60-1 .99 (m, 5H), 1 .83 (m, 2H, J = 8.7 Hz), 1 .56-1 .35 (m, 5H), 1 .28 (d, 6H, J = 4.2 Hz), 1 .09 (d, 3H, J = 1 0.2 Hz), 0.99 (t, 3H, J = 8.7 Hz) ; LC/MS, [M/2+H]+: 635.79, 635.80 calculated.
Step b. Reduction of Nitro-Biafungin To Amino-Biafungin

To a stirring solution of Nitro-Biafungin (1 00 mg, 0.075 mmol) in glacial acetic acid(1 .5 ml_) was added zinc powder (50 mg, 0.77 mmol) and the reaction was stirred at ambient temperature for 1 hour. The mixture was filtered and applied directly to reversed phase HPLC (Isco CombiFlash Rf, 50g Redisep C18 column; 5 to 95% acetonitrile in Dl water containing 0.1 % formic acid: 15 minute gradient). The pure fractions were pooled and lyophilized to yield 55 mg of the desired product as a white solid, formate salt. 1 H-NMR (300 MHz, Methanol-d4) 5 8.47 (bs, 1 H), 7.99 (d, 2H, J = 1 0.8Hz), 7.82 (d, 2H, J = 7.5 Hz), 7.80-7.67 (m, 6H), 7.62 (d, 2H, J = 8.7 Hz), 7.03 (d, 2H, J = 7.5 Hz), 6.77 (d, 1 H, J = 1 .9 Hz), 6.68 (d, 1 H, J = 8.2 Hz), 6.55 (dd, 2H, J = 8.2, 1 .9 Hz), 5.43 (d, 1 H, J = 2.5 Hz), 5.05 (d, 1 H, J = 3 Hz), 4.83-4.73 (m, 2H), 4.64- 4.56 (m, 2H), 4.43-4.34 (m, 2H), 4.31 -4.15 (m, 4H), 4.03-4.08 (m, 1 H), 4.1 1 -3.89 (m, 8H), 3.83 (d, 1 H, J = 1 0.8 Hz), 3.68-3.47 (m, 3H), 3.1 7 (s, 9H), 2.57-2.42 (m, 2H), 2.35-2.27 (m, 1 H), 2.14-1 .98 (m, 2H), 1 .83 (m, 2H, J = 6 Hz), 1 .56-1 .38 (m, 4H), 1 .28 (dd, 6H, J = 6.5, 2 Hz), 1 .09 (d, 3H, J = 7 Hz), 0.986 (t, 3H, J = 7 Hz); High Res LC/MS: [M+H]+ 1241 .61 63; 1241 .6136 calculated.
Step c. Reaction of Amino-Biafungin with lnt-2 to Produce Compound 31

To a stirring solution of Amino-Biafungin (50 mg, 0.04 mmol) in DM F (1 ml_) was added formyl-Met-Leu-Phe- -Ala-OSu (lnt-2) (36 mg, 0.06 mmol) and DI PEA (7 uL, 0.04 mmol). The reaction was stirred at ambient temperature for 1 8 hours. The mixture was applied directly to reversed phase HPLC (Isco CombiFlash Rf; 50g Redisep C1 8 column; 5 to 95% acetonitrile in Dl water containing 0.1 % formic acid: 15 minute gradient). The pure fractions were pooled and lyophilized to yield 26 mg of a white solid as a formate salt. 1 H-NMR (300 M Hz, Methanol-d4) 5 8.55 (bs, 1 H), 8.44 (t, 1 H, J = 10 Hz), 8.1 8 (d, 1 H, J = 6 Hz), 8.1 1 (s, 1 H), 7.99 (d, 2H, J = 1 0 Hz), 7.84-7.70 (m, 6H), 7.63 (d, 2H, J = 7.8 Hz), 7.32-7.1 9 (m, 6H), 7.03 (d, 4H, J = 9 Hz), 6.87 (d, 1 H, J = 8.1 Hz), 5.44 (d, 1 H, J = 1 0.5 Hz), 5.05 (d, 1 H, J = 4.5 Hz), 4.83-4.74 (m, 2H), 4.66-4.50 (m, 6H), 4.45-4.29 (m, 10H), 4.1 9-3.82 (m, 1 0H), 3.67-3.57 (m, 6H), 3.1 7 (s, 9H), 2.64-2.46 (m, 6 H), 2.14-1 .92 (m, 6H), 1 .84 (m, 4H, J = 6 Hz), 1 .62-1 .40 (m, 8H), 1 .32-1 .22 (m, 6H), 1 .09 (d, 3H, J = 9 Hz), 0.99 (t, 3H, J = 7.5 Hz), 0.88 (m, 6H, J = 6.8 Hz) ; High Res LC/MS, [M/2+H]+ 865.4143, 865.4147 calculated.
17 Eschenauer, G; Depestel, DD; Carver, PL (March 2007). “Comparison of echinocandin antifungals.”. Therapeutics and clinical risk management 3 (1): 71–97. PMC 1936290.PMID 18360617.
///////////Biafungin™, CD 101 IV, CD 101 Topical, CD101, SP 3025, PHASE 2, CIDARA, Orphan Drug, Fast Track Designation, Seachaid Pharmaceuticals, Qualified Infectious Disease Product, QIDP, UNII-G013B5478J, 1396640-59-7, 1631754-41-0, Vulvovaginal candidiasis, Echinocandin B, FUNGIN
FREE FORM
CCCCCOc1ccc(cc1)c2ccc(cc2)c3ccc(cc3)C(=O)N[C@H]4C[C@@H](O)[C@H](NC(=O)[C@@H]5[C@@H](O)[C@@H](C)CN5C(=O)[C@@H](NC(=O)C(NC(=O)[C@@H]6C[C@@H](O)CN6C(=O)C(NC4=O)[C@@H](C)O)[C@H](O)[C@@H](O)c7ccc(O)cc7)[C@@H](C)O)OCC[N+](C)(C)C
AND OF ACETATE
CCCCCOc1ccc(cc1)c2ccc(cc2)c3ccc(cc3)C(=O)N[C@H]4C[C@@H](O)[C@H](NC(=O)[C@@H]5[C@@H](O)[C@@H](C)CN5C(=O)[C@@H](NC(=O)C(NC(=O)[C@@H]6C[C@@H](O)CN6C(=O)[C@@H](NC4=O)[C@@H](C)O)[C@H](O)[C@@H](O)c7ccc(O)cc7)[C@@H](C)O)OCC[N+](C)(C)C.CC(=O)[O-]
Three antifungal drugs approved by the United States Food and Drug Administration, caspofungin, anidulafungin, and micafungin, are known to inhibit β-1 ,3-glucan synthase which have the structures shown below.

caspofungin

Anidulafungin

Other exemplary p-1 ,3-glucan synthase inhibitors include,

echinocandin B

cilofungin

pneumocandin A0

pneumocandin B0

L-705589

L-733560

A-174591 

 or a salt thereof,
Biafungin

or a salt thereof,
Amino-biafungin

or a salt thereof,

Amino-AF-053 


ASP9726
Yet other exemplary p-1 ,3-glucan synthase inhibitors include, without limitation:

Papulacandin B


Ergokonin
//////////////

Varenicline (Chantix™)
Varenicline (trade name Chantix and Champix usually in the form of varenicline tartrate), is a prescription medication used to treatnicotine addiction. Varenicline is a nicotinic receptor partial agonist—it stimulates nicotine receptors more weakly than nicotine itself does. In this respect it is similar to cytisine and different from the nicotinic antagonist, bupropion, and nicotine replacement therapies(NRTs) like nicotine patches and nicotine gum. As a partial agonist it both reduces cravings for and decreases the pleasurable effects of cigarettes and other tobacco products. Through these mechanisms it can assist some patients to quit smoking.


 C4H6O6 : 361.35
Varenicline is used for smoking cessation. In a 2009 meta-analysis varenicline was found to be more effective than bupropion (odds ratio 1.40) and NRTs (odds ratio 1.56).[1]
A 2013 Cochrane overview and network meta-analysis concluded that varenicline is the most effective medication for tobacco cessation and that smokers were nearly three times more likely to quit on varenicline than with placebo treatment. Varenicline was more efficacious than bupropion or NRT and as effective as combination NRT for tobacco smoking cessation.[2][3]
The United States’ Food and Drug Administration (US FDA) has approved the use of varenicline for up to twelve weeks. If smoking cessation has been achieved it may be continued for another twelve weeks.[4]
Varenicline has not been tested in those under 18 years old or pregnant women and therefore is not recommended for use by these groups. Varenicline is considered a class C pregnancy drug, as animal studies have shown no increased risk of congenital anomalies, however, no data from human studies is available.[5] An observational study is currently being conducted assessing for malformations related to varenicline exposure, but has no results yet.[6] An alternate drug is preferred for smoking cessation during breastfeeding due to lack of information and based on the animal studies on nicotine.[7]

Varenicline L-tartrate (Compound I) is the international commonly accepted name for 7,8,9,10- tetrahydro-6, 10-methano-6i7-pyrazino [2, 3- h] [3 ] benzazepme, (2R, 3R) -2 , 3-dihydroxybutanedioate (1:1) (which is also known as 5,8,14- tπazatetracyclo [10.3.1. O2‘11. O4‘9] -hexadeca-2 (11) , 3, 5, 7, 9-pentaene, (2R, 3R)-2,3- dihydroxybutanedioate (1:1)) and has an empirical formula of C13H13N3 • C4H6O6 and a molecular weight of 361.35. Varenicline L-tartrate is a commercially marketed pharmaceutically active substance known to be useful for the treatment of smoking addiction.

(D
Varenicline L-tartrate is a partial agonist selective for (X4β2 nicotinic acetylcholine receptor subtypes. In the United States, varenicline L-tartrate is marketed under the name Chantix™ for the treatment of smoking cessation. Varenicline base and its pharmaceutically acceptable acid addition salts are described in U.S. Patent No. 6,410,550. In particular, Example 26 of U.S. Patent No. 6,410,550 describes the preparation of varenicline hydrochloride salt using 1- (4 , 5-dinitro-10- aza-tπcyclo [6.3.1.O2‘7] dodeca-2, 4, 6-trien-10-yl) -2,2,2- tπfluoroethanone (compound of formula (III)) as starting compound. On the other hand, Example HA) of U.S. Patent No. 6,410,550 illustrates the preparation of compound of formula (III) via nitration of compound of formula (II) using an excess of nitronium triflate (>4 equiv) as a nitrating agent. The process disclosed in U.S. Patent No. 6,410,550 is depicted in Scheme 1.

VareniclineΗCl
Scheme 1
However, Coe et al., J. Med. Chem., 48, 3474 (2005), describes the same process and examples as U.S. Patent No. 6,410,550, and it also reveals that this process affords intermediate ortho-4 , 5-dinitrocompound of formula (III) together with the meta-3, 5-dinitro- isomer (i.e. the meta-dinitrocompound) in a ratio 9:1. The presence of the meta-dinitrocompound may affect not only the purity of the intermediate compound of formula III but it may also have an effect on the purity of the final varenicline tartrate, given that it can be carried along the synthetic pathway and/or it can also give rise to other derivative impurities. Thereby, as well as in U.S. Patent No. 6,410,550, in order to isolate pure compound of formula (III) , the raw product is triturated with ethyl acetate/hexane to afford compound of formula (III) with 77% yield. Additionally, the mother liquor is purified by chromatography on silica gel to improve the yield to a total of 82.8%. However, this process is not desirable for industrial implementation since it requires extensive and complicated purification procedures, i.e. trituration of the solid product along with column chromatography purification of the mother liquor, which is not very efficient or suitable for industrial scale-up.
Several improved processes for the synthesis of varenicline or its salts have been reported in the literature (e.g. WO2006/090236) . However, none of these processes tackle the optimization of the purification step of compound of formula (III).
There is therefore the need for providing an improved process for the preparation of varenicline L- tartrate which involves simple experimental procedures well suited to industrial production, which avoids the use of column chromatography purifications, and which affords high pure varenicline L-tartrate which hence can be used directly as a starting product for the preparation of the marketed pharmaceutical speciality.
Additionally, it has been observed that varenicline L-tartrate is usually obtained as a yellow solid under – A –
standard synthetic conditions. In this regard, colour must be attributed to the presence of some specific impurities that may or may not be detectable by conventional methods such as HPLC. The presence of impurities may adversely affect the safety and shelf life of formulations. In this connection, International application No. WO2006/090236 describes the isolation of vareniclme L- tartrate as a white solid. However, in order to remove coloured impurities, the varenicline L-tartrate obtained in WO2006/090236 is treated with a particular activated carbon having a specific grade (i.e. Darco KB-B™) . In fact, Example 5 of WO2006/090236 describes a large reprocessing step which comprises: dissolving varenicline L-tartrate in water, adding toluene, basifying with NaOH aqueous solution, collecting the toluene phase containing varenicline free base, distilling, adding methanol, azeotropically distilling the mixture, and adding more methanol to obtain a methanolic solution containing varenicline free base, adding Darco KB-B™ (10% w/w) , stirring for one hour, filtering through a pad of celite, and treating with L-tartaric acid to give varenicline L- tartrate salt as a white solid. Further, WO2006/090236 provides the absorbance at 430 nm of a varenicline L- tartrate salt solution, either in dichloromethane or in toluene, with or without using Darco KB-B™ activated carbon. However, this measure cannot be used to corroborate the whiteness of the solid varenicline L- tartrate. In addition, Example 3 of International application No. WO2002/092089, also disclose the preparation of varenicline L-tartrate polymorphic form C (i.e. a hydrate polymorph) as a white precipitate. Therefore, there is also a need for a simple and efficient method for preparing varenicline L-tartrate with enhanced whiteness and having a high purity.


Synthesis of Intermediate VIII
J. Med. Chem. 48, 3474 (2005).
http://pubs.acs.org/doi/pdf/10.1021/jm050069n
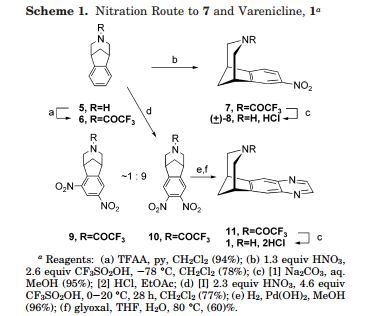
https://www.google.com/patents/WO2001062736A1?cl=en
edited by Harry G. Brittain
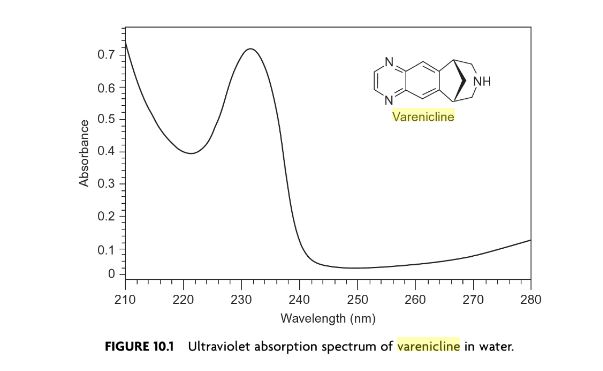
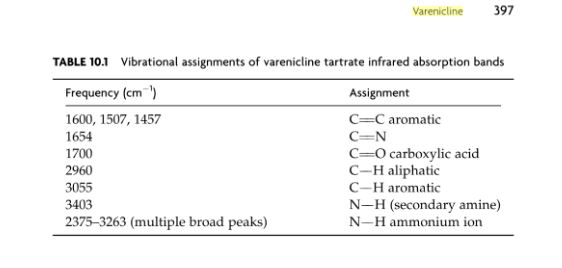
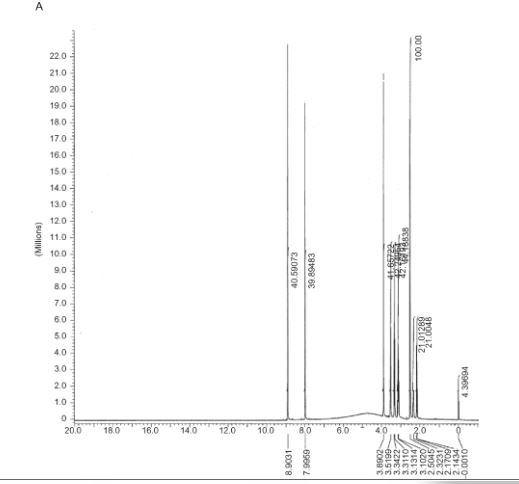

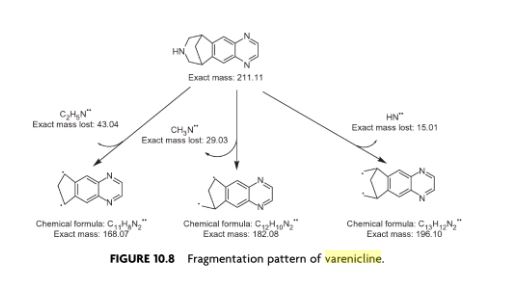

|
DOI: 10.1021/jm00190a020
|
|
| DOI: 10.1021/jm050069n |
CLIP
Scheme (I) compound patent US6410550B1 is provided adjacent difluorobromobenzene as raw materials by DA reaction, oxidation, cyclization, debenzylation get varenicline intermediate (II). The synthesis route is as follows:

Patent CN101693712A mainly given varenicline intermediate (II) The preparation process is different from the compound patented. After the five-step method patents cited compounds. The entire route is longer, while using a large number of precious metal catalysts and reaction conditions need very strict control, inappropriate EVAL industry production.

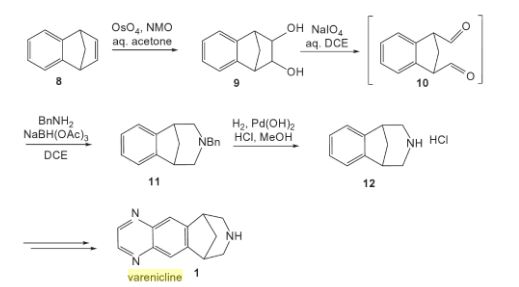
A varenicline intermediate 2,3, 4, 5-tetrahydro-1,5-methylene bridge synthesis -1H-3- benzazepine hydrochloride, which comprises the following Step: (1) 2-indanone of formula 3 and the compound and paraformaldehyde under alkaline or acidic conditions Mannich reaction, as shown in general formula 2 intermediate; (2) the step (I) obtained through reaction of Formula 2 intermediate under basic or acidic conditions by reducing the role of the carbonyl group is reduced to a methylene group, and get varenicline intermediate (II) by debenzylation, the reaction is:

Wherein, R groups are selected from _H, _Me, _Et, _iPr> _t_Bu.
Figure 2;

Wherein, R group is -H, -Me, -Et, -iPr or -t_Bu.
(2) Step (I) obtained by the reaction intermediates of formula under basic or acidic conditions by reducing the role of the carbonyl group is reduced 2 methylene, and get by debenzylation cutting Lenk Lin intermediate (II);

Varenicline, a nicotinic 42 partial agonist, was approved in the US for the treatment of smoking cessation in May of 2006. It was developed and marketed by Pfizer as a treatment for cigarette smokers who want to quit. Varenicline partially activates the nicotinic receptors and thus reduces the craving for cigarette that smokers feel when they try to quit smoking. By mitigating this craving and antagonizing nicotine activity without other symptoms, this novel drug helps quitting this dangerous addiction easier on the patients [6,52]. Several modifications [54,55] to the original synthesis [53,56] have been reported in the literature, including an improved process scale synthesis of the last few steps (Scheme 15) [57]. The Grignard reaction was initiated on a small scale by addition of 2-bromo fluorobenzene 113 to a slurry of Magnesium turnings and catalytic 1,2-dibromoethane in THF and heating the mixture until refluxing in maintained. To this refluxing mixture was added a mixture of the 2-bromo fluorobenzene 113 and cyclopentadiene 114 over a period of 1.5 h. After complete addition, the reaction was allowed to reflux for additional 1.5 h to give the Diels- Alder product 115 in 64% yield. Dihydroxylation of the olefin 115 by reacting with catalytic osmium tetraoxide in the presence of N-methylmorpholine N-oxide (NMO) in acetone: water mixture at room temperature provided the diol 116 in 89% yield. Oxidative cleavage of diol 116 with sodium periodate in biphasic mixture of water: DCE at 10ºC provided di-aldehyde 117 which was immediately reacted with benzyl amine in the presence of sodium acetoxyborohydride to give benzyl amine 118 in 85.7% yield. The removal of the benzyl group was effected by hydrogenation of the HCl salt in 40-50 psi hydrogen pressure with 20% Pd(OH)2 in methanol to give amine hydrochloride 119 in 88% yield. Treatment of amine 119 with trifluoroacetic anhydride and pyridine in dichloromethane at 0ºC gave trifluoroacetamide 120 in 94% yield. Dinitro compound 121 was prepared by addition of trifluoroacetamide 120 to a mixture of trifluoromethane sulfonic acid and nitric acid, which was premixed, in dichloromethane at 0ºC. Reduction of the dinitro compound 121 by hydrogenation at 40-50 psi hydrogen in the presence of catalytic 5%Pd/C in isopropanol:water mixture provided the diamine intermediate 122 which was quickly reacted with glyoxal in water at room temperature for 18h to give compound 123 in 85% overall yield. The trifluoroacetamide 123 was then hydrolyzed with 2 M sodium hydroxide in toluene at 37-40ºC for 2-3h followed by preparation of tartrate salt in methanol to furnish varenicline tartrate (XV).
[52]Keating, G.; Siddiqui, M. A. A. CNSdrugs, 2006, 11, 946.
[53] Coe, J. W.; Brooks, P. R.; Vetelino, M. G.; Wirtz, M. C.; Arnold,E. P. ; Huang, J.; Sands, S. B.; Davis, T. I.; Lebel, L. A.; Fox, C.
B.; Shrikhande, A.; Heym, J. H.; Schaeffer, E.; Rollema, H.; Lu,Y.; Mansbach, R. S.; Chambers, L. K.; Rovetti, C. C.; Schulz, D.
W.; Tingley, III, F. D.; O’Neill, B. T. J. Med. Chem., 2005, 48,3474.
[54] Brooks, P. R.; Caron, S.; Coe, J. W.; Ng, K. K.; Singer, R. A.;Vazquez, E.; Vetelino, M. G.; Watson, Jr. H. H.; Whritenour, D.
C.; Wirtz, M. C. Synthesis, 2004, 11, 1755.
[55] Singer, R. A.; McKinley, J. D.; Barbe, G.; Farlow, R. A. Org. Lett.,2004, 6, 2357.
[56] Coe, J. W.; Brooks, P. R. P. US-6410550 B1, 2002.
[57] Busch, F. R.; Hawkins, J. M.; Mustakis, L. G.; Sinay, T. G., Jr.;Watson, T. J. N.; Withbroe, G. J. WO-2006090236 A1, 2006.
WO 2002085843
https://google.com/patents/WO2002085843A2?cl=en
https://www.google.com/patents/EP2204369A1?cl=en
Varenicline (a compound I of formula I) is the international commonly accepted non-proprietary name for 7,8,9,10-tetrahydro-6,10-methano-6H-pyrazino[2,3-h][3]benzazepine (which is also known as 5,8,14-triazatetracyclo[10.3.1.02,11.04,9]-hexadeca-2(11),3,5,7,9-pentaene), and has an empirical formula of C13H13N3 and a molecular weight of 211.26.

The L-tartrate salt of varenicline is known to be therapeutically useful and is commercially marketed for the treatment of smoking addiction. Varenicline L-tartrate is a partial agonist selective for α4β2 nicotinic acetylcholine receptor subtypes. In the United States, varenicline L-tartrate is marketed under the trade mark Chantix and is indicated as an aid to smoking cessation treatment.
Varenicline base and its pharmaceutically acceptable acid addition salts are described in U.S. Patent No. 6,410,550 . In particular, the preparation of varenicline provided in this reference makes use of 10-aza-tricyclo[6.3.1.02,7]-dodeca-2(7),3,5-triene (a compound of Formula VI), as a key intermediate compound (see Scheme 1 below). Specifically, Example 1 of U.S. Patent No. 6,410,550 describes the synthetic preparation of key intermediate compound of Formula VI as depicted in Scheme 1.

1,2,3,4-tetrahydro-1,4-methano-naphthalene-cis-2,3-diol (a compound of Formula III), and / or indane-1,3-dicarbaldehyde (a compound of Formula IV).
Example 1: Preparation of 1,2,3,4-tetrahydro-1,4-methano-naphthalene-cis-2,3-diol (a compound of Formula III)
A 10mL round bottom flask was charged with a compound of formula II (142mg, 1mmol), N-methylmorpholine-N-oxide (120mg, 1.03mmol), tert-butanol (3mL) and water (1mL). FibreCat™ 3003 (OsO4 anchored onto a polymeric support) (11.6mg, 0.0025mmol) was added to this solution and the mixture was heated to reflux. Complete conversion to a compound of formula III was detected by GC, method A, after 48h.
Example 2: Preparation of 1,2,3,4-tetrahydro-1,4-methano-naphthalene-cis-2,3-diol (a compound of Formula III)Step A) Preparation of hexadecyl-trimethylammoniumpermanganate (HTAP):
HTAP was prepared from ion exchange reaction between hexadecyltrimethylammoniumbromide and potassium permanganate.
Potassium permanganate (17.38g, 0.11mol, 1equiv.) was dissolved in 500mL water. A solution of hexadecyltrimethylammoniumbromide (40.10g, 0.11mol, 1equiv) in 500mL water was added drop-wise over 45 min at 20-22°C, and the mixture stirred for 30 minutes at this temperature. The precipitated solid was collected by filtration, washed with water (3 x 100mL) and dried under vacuum at 35°C for 24 hours to give 34.38g of HTAP as a light purple solid.
Step B) Preparation of a compound of formula III:
Compound II (3.52g, 24.8mmol, 1equiv.) was dissolved in anhydrous tetrahydrofuran (80mL) and a solution of HTAP (10g, 24.8mmol, 1.0equiv.) in anhydrous tetrahydrofuran (125mL) was added drop-wise at 23-30°C over 45min. The reaction was monitored by TLC (hexane-ethyl acetate = 1:1). After complete reaction the mixture was cooled to below 10°C, and methyl tert-butyl ether (50mL) and 5% aqueous NaOH solution (50mL) were added and the mixture stirred for 30min. The solid was removed by filtration, and washed with methyl tert-butyl ether (2 x 30mL). The combined layers of the filtrate were separated and the aqueous phase extracted with methyl tert-butyl ether (2 x 30mL). The organic layers were combined and washed with 5% aqueous NaOH solution (50mL), water (2 x 50mL), dried over MgSO4, filtered and concentrated to obtain a dark green solid. This residue was suspended in acetone (15mL) and collected by filtration, washing with additional acetone (3 x 5mL). The product was dried under vacuum at 40°C to give 2.215g (50.7% yield) as a white crystalline solid.
Analytical data: m.p. = 178.8-179.3°C; 1H-NMR: See Figure 1; 13C-NMR: See Figure 2.
Example 3: Preparation of indane-1,3-dicarbaldehyde (a compound of Formula IV)
A 25 mL round bottom flask was charged with a compound of formula I (142mg, 1mmol), Ruthenium (III) chloride hydrate (Aldrich, Reagent Plus™) (7.2mg, 0.035mmol), acetonitrile (8.5mL) and water (1.1mL). The solution was heated to 45°C and sodium periodate (449mg, 2.1mmol) was added portionwise over 25 minutes. After 1h, the reaction was cooled to ambient temperature and filtered. The solids were washed with ethyl acetate (3 x 2mL) and water (3mL). The filtrate was concentrated under vacuum and 5mL of water were added to the obtained residue. The mixture was extracted with ethyl acetate (2 x 5mL) and the combination of the organic layers was washed with water (3 x 5mL), dried with MgSO4 and concentrated under vacuum to obtain a compound of formula IV (118mg) in 68% yield, 70.9% purity (analyzed by GC, method A).
WO 199935131, WO 2002092089, US 2013030179
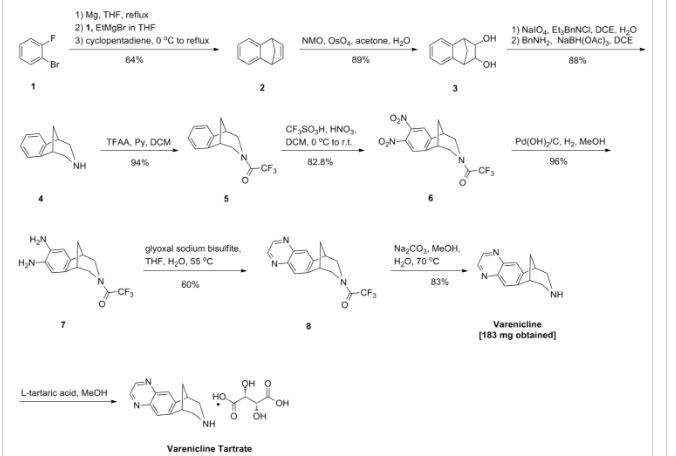
https://www.google.com/patents/WO2009065872A2?cl=en
Example 1: Preparation of 7,8,9,10- tetrahydro-6, 10-methano-6H-pyrazino [2, 3-h] [3] benzazepine L-tartrate (i.e. varenicline L-tartrate)
A) Preparation of compound of formula (III)
This example is based on U.S. Patent No. 6,410,550.
A 250 mL round bottom flask with thermometer, condenser, addition funnel and magnetic stirring was charged with 10-aza-tricyclo [ 6.3.1. O2‘7] dodeca-2, 4, 6- triene para-toluene sulfonic acid salt (12.4g, 37.5 mmol) and 44 mL of CH2Cl2. Triethylamine (8.3 g, 82.5 mmol) was added to the slurry and the resulting solution was cooled to 0-5 0C. The addition funnel was charged with a solution of (CF3CO)2O (8.1q, 41.25 mmol) in 19 mL of CH2Cl2. This solution was slowly added to the reaction mixture, maintaining the temperature < 15 0C. The resulting mixture was stirred for 1 hour, and the complete conversion was monitored by GC. The crude reaction mixture was washed with water (2 * 40 mL) and brine (40 mL) . The organic phase was used in the next step without further purification.
On the other hand, a 500 mL round bottom flask with thermometer, condenser, addition funnel and magnetic stirring was charged with CF3SO3H (25.9 g, 172.5 mmol), CH2Cl2 (110 mL) and cooled to 0-5 0C. At this temperature, fuming nitric acid (5.4 g, 86.25 mmol) was added slowly. To the resulting slurry at 0-5 0C, the solution obtained in the previous step was slowly added, maintaining the temperature < 15 0C. After the addition, the reaction mixture was stirred overnight. The complete dinitration was confirmed by GC. The crude reaction mixture was poured into water (60 mL) an ice (80 g) and stirred. The phases were separated and the aqueous phase was extracted with CH2Cl2 (3 x 50 mL) . The mixture of the organic phases was washed with aqueous saturated NaHCO3, dried over Na2SO4 and volatiles evaporated under vacuum to obtain 11.9 g of a solid that was suspended and stirred for 2 hours in AcOEt (12 mL) and hexanes (24 mL) . The solid was filtered and washed with hexanes to obtain the compound of formula (III), 9.1g with a purity of 88.9% by GC (9.8% of meta-dimtrocompound impurity) .
B) Preparation of compound of formula (IV)
This example is based on International Patent No. WO/2006/090236.
A 200 mL autoclave was charged with (III) (9.1 g, 26.3 mmol), damp 5% Pd/C 50% and 180 mL of a 2- propanol/water (80/20 wt/wt) . The reaction was stirred under 50 psi of hydrogen for 18 hours. The complete hydrogenation was confirmed by GC analysis. The reaction was filtered through Celite and washed with 2-propanol (40 mL) . To this solution, K2HPO4(458 mg, 2.63 mmol) was added. The mixture was cooled at 0-5 0C and a solution of 4.07 g of 40% aqueous glyoxal diluted with water (14.5 mL) was added slowly. The resulting solution was stirred 2 hours at this temperature and overnight at room temperature. The complete conversion was confirmed by GC analysis. The reaction was concentrated under vacuum to a volume of 68 mL and water (128 mL) was added drop- wise. The resulting suspension was stirred for 2 hours at room temperature, 1 hour in a ice/water bath, filtered, washed with water (20 mL) and dried m a oven at 50 0C to obtain the compound of formula (IV), 6.78 g.
C) Preparation of vareniclme L-tartrate (compound of formula (I) )
This example is based on International Patent No. WO/2006/090236.
A 250 mL round bottom flask with thermometer, condenser, and magnetic stirring was charged with compound of formula (IV) (6.78 g, 22 mmol) and toluene
(47 mL) . To this solution was added a solution of NaOH (2.7 g, 68.2 mmol) in water (34 mL) . The mixture was heated to 400C and stirred for 4 hours. The complete hydrolysis was confirmed by GC analysis. Toluene (68 mL) was added and the reaction was cooled. The phases were separated and the aqueous phase was extracted with toluene (30 mL) . The organic phases were evaporated under vacuum. The residue was dissolved in MeOH (90 mL) and evaporated again. The final residue was dissolved in 156 mL of MeOH. 1.3 g of activated carbon “Darco G-60 100 mesh” were added and the mixture was stirred for 30 min and filtered through Celite to obtain an intense yellow solution. The process with activated carbon was repeated without any improvement in the colour. This solution was added drop-wise over a solution of L- tartaric acid (3.63 g, 24.2 mmol) in MeOH (47 mL) . The slurry was stirred for 72 hours at room temperature, filtered, washed with MeOH and dried in an oven at 50 0C for 8 hours, to obtain 5.05 g of varenicline L-tartrate as a yellow solid with a 95.5% purity by HPLC (4.4% of unknown impurity A). Colour L: 92.75, a*: -7.19, b*:43.08.
Comparative Example 2: Preparation of 7,8,9,10- tetrahydro-6, 10-methano-6H-pyrazmo [2, 3-h] [3 ] benzazepine L-tartrate (i.e. varenicline L-tartrate) A) Preparation of compound of formula (IV)
This example is based on International Patent No. WO/2006/090236.
A 200 mL autoclave was charged with (III) prepared according to Comparative Example 1.A) (4.1 g) , 123 mg of damp 5% Pd/C 50% and 81 mL of a 2-propanol/water (80/20 wt/wt) . The reaction was stirred under 50 psi of hydrogen for 24 hours. The complete hydrogenation was confirmed by GC analysis. The reaction was filtered through Celite and washed with 2-propanol (16 mL) . To this solution, K2HPO4 (207 mg, 1.19 mmol) was added. The mixture was cooled at 0-5 0C and a solution of 1.84 g of 40% aqueous glyoxal diluted with water (6.6 mL) was added slowly. The resulting solution was stirred 2 hours at this temperature and overnight at room temperature. The complete conversion was confirmed by GC analysis. The reaction was concentrated under vacuum to a volume of 30 mL and water (56 mL) was added drop-wise. The resulting suspension was stirred for 2 hours at room temperature, 1 hour in a ice/water bath, filtered, washed with water and dried in a oven at 50 0C to obtain 3.15 g of compound of formula (IV) .
B) Preparation of vareniclme L-tartrate (compound of formula (I) )
This example is based on International application No. WO/2006/090236. A 100 mL round bottom flask with thermometer, condenser, and magnetic stirring was charged with
7, 8, 9, 10-tetrahydro-8- (tπfluoroacetyl) -6, 10-methano-6H- pyrazino [2 , 3-h] [3] benzazepine, i.e. compound of formula
(IV) (3.14 g, 10.2 mmol) and toluene (22 mL) . To this solution was added a solution of NaOH (1.3 g, 31.6 mmol) in water (16 mL) . The mixture was heated to 40 0C and stirred for 2.5 hours. The complete hydrolysis was confirmed by GC analysis. Toluene (30 mL) was added and the reaction was cooled. The phases were separated and the aqueous phase was extracted with toluene (15 mL) . The organic phases were evaporated under vacuum. The residue was dissolved in MeOH (45 mL) and evaporated again. The final residue was dissolved m 70 mL of MeOH. 314 mg of activated carbon “Darco G-60 100 mesh” were added and the mixture was stirred for 30 mm and filtered through Celite to obtain a yellow solution. This solution was added drop-wise over a solution of L- tartaπc acid (1.68 g, 11.22 mmol) m MeOH (22 mL) . The slurry was stirred for 1 hour at room temperature, filtered, washed with MeOH (2 x 5 mL) and dried under vacuum, to obtain vareniclme L-tartrate (2.48 g) as a yellow solid with a 95.6% purity by HPLC (4.4% of unknown impurity A). Colour L: 99.50, a*: -4.98, b*:43.02
Comparative Example 3: Preparation of 7,8,9,10- tetrahydro-6, 10-methano-6H-pyrazino [2, 3-h] [3 ] benzazepine L-tartrate (i.e. vareniclme L-tartrate)
This example is based on International application No. WO/2002/092089.
2 g of vareniclme L-tartrate as obtained from Comparative Example 1 were dissolved in 3 mL of water.
To this solution, 100 mL of CH3CN were added, and the resulting slurry was stirred for 10 mm and filtered.
After drying the product was analysed to be a 98.2% purity by HPLC (1.7% of unknown impurity A) . Colour L: 91.44, a*: -3.24, b* : 33.47
Example 1: Preparation of 7, 8, 9, lO-tetrahydro-6, 10- methano-6H-pyrazmo [2, 3-h] [3] benzazepine L-tartrate
(i.e. vareniclme L-tartrate)
A) Preparation of compound of formula (III) This example is based on U.S. Patent No. 6,410,550, except for the purification step, which is the object of the present invention (i.e. crystallization in toluene) .
A 500 mL round bottom flask with thermometer, condenser, addition funnel and magnetic stirring was charged with 10-aza-tricyclo [ 6.3.1. O2‘7] dodeca-2, 4, 6- tπene para-toluene sulfonic acid salt (32.5g, 98.2 mmol) and 115 mL of CH2Cl2. Triethylamine (21.8 g, 216 mmol) was added to the slurry and the resulting solution was cooled to 0-5 0C. The addition funnel was charged with a solution of (CF3CO)2O (22.7 g, 108 mmol) in 50 mL of CH2Cl2. This solution was slowly added to the reaction mixture, maintaining the temperature < 15 0C. The resulting mixture was stirred for 1 hour, and the complete conversion was monitored by GC. The crude reaction mixture was washed with water (2 x 100 mL) and brine (100 mL) . The organic phase was used in the next step without further purification.
A l L round bottom flask with thermometer, condenser, addition funnel and magnetic stirring was charged with CF3SO3H (67.8 g, 452 mmol), CH2Cl2 (280 mL) and cooled to 0-5 0C. At this temperature, fuming nitric acid (14.2 g, 226 mmol) was slowly added. To the resulting slurry at 0-5 0C, the solution obtained in the previous step was slowly added, maintaining the temperature < 15 0C. After the addition, the reaction mixture was stirred overnight. The complete dinitration was confirmed by GC. The crude reaction mixture was poured into water (150 mL) an ice (200 g) and stirred. The phases were separated and the aqueous phase was extracted with CH2Cl2 (100 mL) . The mixture of the organic phases was washed with aqueous saturated NaHCO3 (2×100 mL) , water (100 mL) , dried over Na2SO4 and volatiles evaporated under vacuum to obtain 30.5 g of a solid with a 83.6% purity by GC (12.5% of meta- dinitrocompound impurity) . 20 g of this solid were crystallized in toluene (100 mL) to obtain the compound of formula (III), 15 g of a pale brown solid with a 98.5 % purity by GC (meta-dinitrocompound impurity not detected) .
B) Preparation of compound of formula (IV) This example is based on International Patent No. WO/2006/090236.
A 200 mL autoclave was charged with (III) (9.1 g, 26.3 mmol, crystals from toluene), damp 5% Pd/C 50% and 180 mL of a 2-propanol/water (80/20 wt/wt) . The reaction was stirred under 50 psi of hydrogen for 18 hours. The complete hydrogenation was confirmed by GC analysis. The reaction was filtered over Celite and washed with 2- propanol (40 mL) . To this solution, K2HPO4 (458 mg, 2.63 mmol) was added. The mixture was cooled at 0-5 0C and a solution of 4.07 g of 40% aqueous glyoxal diluted with water (14.5 mL) was added slowly. The resulting solution was stirred 2 hours at this temperature and overnight at room temperature. The complete conversion was confirmed by GC analysis. The reaction was concentrated under vacuum to a volume of 68 mL and water (128 mL) was added drop-wise. The resulting suspension was stirred for 2 hours at room temperature, 1 hour in a ice/water bath, filtered, washed with water (20 mL) and dried m a oven at 50 0C to obtain the product, 7.16 g of compound of formula (IV) with a 99.9% purity by HPLC. C) Preparation of varenicline L-tartrate (compound of formula ( I) )
Thrs example rs based on International Patent No. WO/2006/090236. A 250 mL round bottom flask with thermometer, condenser, and magnetic stirring was charged with a solution of NaOH (2.89 g, 72.23 mmol) in water (36 mL) , compound of formula (IV) (7.15 g, 23.3 mmol) and toluene (50 mL) . The mixture was heated to 40 0C and stirred for 4 hours. The complete hydrolysis was confirmed by GC analysis. Toluene (71 mL) was added and the reaction was cooled. The phases were separated and the aqueous phase was extracted with toluene (36 mL) . The organic phases were evaporated under vacuum. The residue was dissolved in MeOH (110 mL) and evaporated again. The final residue was dissolved in 164 mL of MeOH. 750 mg of activated carbon “Darco G-60 100 mesh” were added and the mixture was stirred for 30 min and filtered through Celite to obtain a yellow solution. This solution was added drop- wise over a solution of L-tartaric acid (3.84 g, 25.6 mmol) in MeOH (50 mL) . The slurry was stirred for 14 hours at room temperature, filtered, washed with MeOH and dried under vacuum, to obtain varenicline L-tartrate
(7.04 g) as an off-white solid with a >99.9% purity by HPLC (unknown impurity A not detected) . Colour L: 94.39, a*: 2.27, b*:9.02.

No evidence for increased risks of cardiovascular events, depression, or self-harm with varenicline versus nicotine replacement therapy has been found in one post-marketing surveillance study.[23]
Varenicline displays full agonism on α7 nicotinic acetylcholine receptors.[24][25] And it is a partial agonist on the α4β2, α3β4, and α6β2 subtypes.[26] In addition, it is a weak agonist on the α3β2 containing receptors.
Varenicline’s partial agonism on the α4β2 receptors rather than nicotine’s full agonism produces less effect of dopamine release than nicotine’s. This α4β2 competitive binding, reduces the ability of nicotine to bind and stimulate the mesolimbic dopamine system – similar to the method of action of buprenorphine in the treatment of opioid addiction.[3]
Most of the active compound is excreted by the kidneys (92–93%). A small proportion is glucuronidated, oxidised, N-formylated or conjugated to a hexose.[27] The elimination half-life is about 24 hours.
Use of Cytisus plant as a smoking substitute during World War II[28] led to use as a cessation aid in eastern Europe and extraction of cytisine.[29] Cytisine analogs led to varenicline at Pfizer.[30][31][32]
Varenicline received a “priority review” by the US FDA in February 2006, shortening the usual 10-month review period to 6 months because of its demonstrated effectiveness inclinical trials and perceived lack of safety issues.[33] The agency’s approval of the drug came on May 11, 2006.[4] On August 1, 2006, varenicline was made available for sale in the United States and on September 29, 2006, was approved for sale in the European Union.[34]
SEE
| US6410550 | Nov 13, 1998 | Jun 25, 2002 | Pfizer Inc | Aryl fused azapolycyclic compounds |
| WO2009155403A2 * | Jun 18, 2009 | Dec 23, 2009 | Teva Pharmaceutical Industries Ltd. | Processes for the preparation of varenicline and intermediates thereof |
| Reference | ||
|---|---|---|
| 1 | * | BHUSHAN, VIDYA; RATHORE, RAJENDRA; CHANDRASEKARAN, S.: “A Simple and Mild Method for the cis-Hydroxylation of Alkenes with Cetyltrimethylammonium Permanganate” SYNTHESIS, no. 5, 1984, pages 431-433, XP002581198 |
| 2 | * | BROOKS P R ET AL: “Synthesis of 2,3,4,5-tetrahydro-1,5-methano-1H-3-benzaz epine via oxidative cleavage and reductive amination strategies” SYNTHESIS 20040803 DE, no. 11, 3 August 2004 (2004-08-03), pages 1755-1758, XP002581197 ISSN: 0039-7881 |
| 3 | * | SORBERA L A ET AL: “Varenicline tartrate: Aid to smoking cessation nicotinic [alpha]4[beta]2 partial agonist” DRUGS OF THE FUTURE 200602 ES LNKD- DOI:10.1358/DOF.2006.031.02.964028, vol. 31, no. 2, February 2006 (2006-02), pages 117-122, XP002581199 ISSN: 0377-8282 DOI: 10.1358/dof.2006.031.02.964028 |
| WO2001062736A1 * | Feb 8, 2001 | Aug 30, 2001 | Pfizer Products Inc. | Aryl fused azapolycyclic compounds |
| WO2002085843A2 * | Mar 4, 2002 | Oct 31, 2002 | Pfizer Products Inc. | Process for the preparation of 1,3-substituted indenes and aryl-fused azapolycyclic compounds |
| WO2006090236A1 * | Feb 21, 2006 | Aug 31, 2006 | Pfizer Products Inc. | Preparation of high purity substituted quinoxaline |
| WO2008060487A2 * | Nov 9, 2007 | May 22, 2008 | Pfizer Products Inc. | Polymorphs of nicotinic intermediates |
| Reference | ||
|---|---|---|
| 1 | * | COE J W ET AL: “Varenicline: an alpha4beta2 Nicotinic Receptor Partial Agonist for Smoking Cessation” JOURNAL OF MEDICINAL CHEMISTRY, AMERICAN CHEMICAL SOCIETY, WASHINGTON., US, vol. 48, no. 10, 1 January 2005 (2005-01-01), pages 3474-3477, XP002474642 ISSN: 0022-2623 cited in the application |
| Citing Patent | Filing date | Publication date | Applicant | Title |
|---|---|---|---|---|
| WO2010005643A1 * | May 28, 2009 | Jan 14, 2010 | Teva Pharmaceutical Industries Ltd. | Processes for purifying varenicline l-tartrate salt and preparing crystalline forms of varenicline l-tartrate salt |
| WO2011110954A1 * | Mar 8, 2011 | Sep 15, 2011 | Actavis Group Ptc Ehf | Highly pure varenicline or a pharmaceutically acceptable salt thereof substantially free of methylvarenicline impurity |
| WO2011154586A3 * | Jun 13, 2011 | Mar 22, 2012 | Medichem, S. A. | Improved methods for the preparation of quinoxaline derivatives |
| EP2581375A2 * | Jun 13, 2011 | Apr 17, 2013 | Medichem, S.A. | Improved methods for the preparation of quinoxaline derivatives |
| US8039620 | May 21, 2009 | Oct 18, 2011 | Teva Pharmaceutical Industries Ltd. | Varenicline tosylate, an intermediate in the preparation process of varenicline L-tartrate |
| US8178537 | Jun 22, 2010 | May 15, 2012 | Teva Pharmaceutical Industries Ltd. | Solid state forms of varenicline salts and processes for preparation thereof |
|
|
 |
|
| Systematic (IUPAC) name | |
|---|---|
|
7,8,9,10-Tetrahydro-6,10-methano-6H-pyrazino[2,3-h] [3]benzazepine
|
|
| Clinical data | |
| Trade names | Chantix |
| AHFS/Drugs.com | Monograph |
| MedlinePlus | a606024 |
| License data |
|
| Pregnancy category |
|
| Routes of administration |
Oral |
| Legal status | |
| Legal status | |
| Pharmacokinetic data | |
| Protein binding | <20% |
| Metabolism | Limited (<10%) |
| Biological half-life | 24 hours |
| Excretion | Renal (81–92%) |
| Identifiers | |
| CAS Number | 249296-44-4 375815-87-5 |
| ATC code | N07BA03 (WHO) |
| PubChem | CID 5310966 |
| IUPHAR/BPS | 5459 |
| DrugBank | DB01273 |
| ChemSpider | 4470510 |
| UNII | W6HS99O8ZO |
| KEGG | D08669 |
| ChEBI | CHEBI:84500 |
| ChEMBL | CHEMBL1076903 |
| Chemical data | |
| Formula | C13H13N3 |
| Molar mass | 211.267 g/mol |
////////////Varenicline, Chantix™, FDA 2006, 249296-44-4, 375815-87-5, Champix , Pfizer, バレニクリン酒石酸塩
n1c2cc3c(cc2ncc1)[C@@H]4CNC[C@H]3C4
What needs to be considered if an API is produced in the time period of a valid written confirmation but imported after this confirmation has expired? This is answered in a revised Q&A Document of the EU Commission.
The EU Commission has updated its Question and Answers Document “Importation of active substances for medicinal products for human use” (now version 7). In this updated version, the question “Can an API batch manufactured during the period of validity of a written confirmation be imported into the EU once the written confirmation is expired?”
In the answer it is referred to Article 46(b)(2)(b) of Directive 2001/83/EC, where it is defined that APIs can only be imported if they are manufactured in accordance with EU GMP or equivalent, and accompanied by a written confirmation from the competent authority of the exporting third country certifying this.
But what if an…
View original post 167 more words

In January and March 2015, the U.K. Medicines and Healthcare Products Regulatory Agency (MHRA) published a “GMP Data Integrity Definitions and Guidance for Industry”. The agency has recently published a new version of the Guidance. Please find here a short overview of the new features in the “GxP Data Integrity Definitions and Guidance for Industry: Draft version for consultation”.
In recent years, regulatory authorities have been struggling with data integrity issues. In particular the U.S. American FDA has tightened the awareness regarding the topic in many Warning Letters. In the meantime, data integrity has also become a focus of European regulatory authorities’ inspections. One of the first regulatory authorities to publish a “GMP Data Integrity Definitions and Guidance for Industry” in January and March 2015 was the U.K. Medicines and Healthcare Products Regulatory Agency (MHRA). More information can be found in “MHRA revises its Guideline on Data Integrity in the short Term
View original post 172 more words