PROUD Indian WORLD RECORD VIEWS holder on THIS BLOG, ………live, by DR ANTHONY MELVIN CRASTO, Worldpeaceambassador, Worlddrugtracker, Helping millions, 100 million hits on google, pushing boundaries,2.5 lakh plus connections worldwide, 45 lakh plus VIEWS on this blog in 227 countries, 7 CONTINENTS ……A 90 % paralysed man in action for you, I am suffering from transverse mylitis and bound to a wheel chair, [THIS BLOG HOLDS WORLD RECORD VIEWS ] A PROUD INDIAN
DR ANTHONY MELVIN CRASTO, Born in Mumbai in 1964 and graduated from Mumbai University, Completed his Ph.D from ICT, 1991,Matunga, Mumbai, India, in Organic Chemistry, The thesis topic was Synthesis of Novel Pyrethroid Analogues, Currently he is working with AFRICURE PHARMA, ROW2TECH, NIPER-G, Department of Pharmaceuticals, Ministry of Chemicals and Fertilizers, Govt. of India as ADVISOR, earlier assignment was
with GLENMARK LIFE SCIENCES LTD, as CONSUlTANT, Retired from GLENMARK in Jan2022 Research Centre as Principal Scientist, Process Research (bulk actives) at Mahape, Navi Mumbai, India. Total Industry exp 32 plus yrs, Prior to joining Glenmark, he has worked with major multinationals like Hoechst Marion Roussel, now Sanofi, Searle India Ltd, now RPG lifesciences, etc. He has worked with notable scientists like Dr K Nagarajan, Dr Ralph Stapel, Prof S Seshadri, etc, He did custom synthesis for major multinationals in his career like BASF, Novartis, Sanofi, etc., He has worked in Discovery, Natural products, Bulk drugs, Generics, Intermediates, Fine chemicals, Neutraceuticals, GMP, Scaleups, etc, he is now helping millions, has 9 million plus hits on Google on all Organic chemistry websites. His friends call him Open superstar worlddrugtracker. His New Drug Approvals, Green Chemistry International, All about drugs, Eurekamoments, Organic spectroscopy international,
etc in organic chemistry are some most read blogs He has hands on experience in initiation and developing novel routes for drug molecules
and implementation them on commercial scale over a 32 PLUS year tenure till date Feb 2023, Around 35 plus products in his career. He has good knowledge of IPM, GMP, Regulatory aspects, he has several International patents published worldwide . He has good proficiency in Technology transfer, Spectroscopy, Stereochemistry, Synthesis, Polymorphism etc., He suffered a paralytic stroke/ Acute Transverse mylitis in Dec 2007 and is 90 %Paralysed, He is bound to a wheelchair, this seems to have injected feul in him to help chemists all around the world, he is more active than before and is pushing boundaries, He has 100 million plus hits on Google, 2.5 lakh plus connections on all networking sites, 100 Lakh plus views on dozen plus blogs, 227 countries, 7 continents, He makes himself available to all, contact him on +91 9323115463, email amcrasto@gmail.com, Twitter, @amcrasto , He lives and will die for his family, 90% paralysis cannot kill his soul., Notably he has 38 lakh plus views on New Drug Approvals Blog in 227 countries......https://newdrugapprovals.wordpress.com/ , He appreciates the help he gets from one and all, Friends, Family, Glenmark, Readers, Wellwishers, Doctors, Drug authorities, His Contacts, Physiotherapist, etc
He has total of 32 International and Indian awards
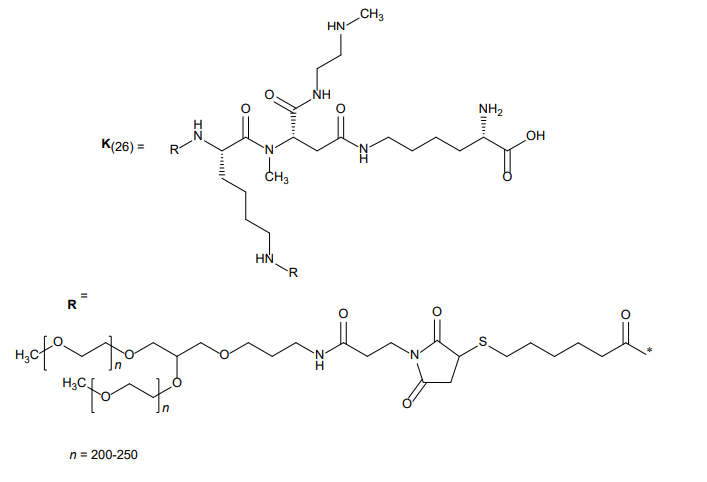
MOLECULAR FORMULA C231H386N64O67S5 + (C2H4O)4n MOLECULAR WEIGHT approx. 45 kDa
The structure of navepegritide (YUVIWEL®) is built using a “prodrug” design. It is not a simple small molecule, but rather a complex conjugate consisting of three distinct components designed to release the active drug slowly over time.
1. The Active Part: C-Type Natriuretic Peptide (CNP)
The core of the molecule is a synthetic 38-amino acid peptide (CNP-38).
Sequence: This peptide mimics the natural human C-type natriuretic peptide, which is essential for bone growth.
Function: Once released, this peptide binds to the natriuretic peptide receptor B (NPR-B) on the surface of chondrocytes (cartilage cells) in the growth plates, stimulating bone formation.
2. The Carrier: Polyethylene Glycol (PEG)
To prevent the body from clearing the small peptide too quickly, it is attached to a large, inert carrier.
Type: It uses a multi-arm, branched 40 kDa Polyethylene Glycol (PEG) molecule.
Purpose: The PEG carrier acts as a shield and a “weight,” making the molecule too large to be filtered out rapidly by the kidneys. This is what allows for once-weekly dosing instead of daily injections.
3. The Linker: TransCon™ Technology
This is the most critical part of the structure. The peptide is attached to the PEG carrier via a cleavable linker.
Mechanism: This linker is designed to break down spontaneously at a predictable rate under physiological conditions (neutral pH and body temperature).
The Result: As the linker slowly breaks, it releases the unmodified, active CNP-38 into the bloodstream. Because the peptide is released in its natural state, it retains its full biological activity.
Summary Table: Structural Components
Component
Description
Role
Peptide
CNP-38 (38 amino acids)
The “payload” that stimulates bone growth.
Linker
pH-sensitive cleavable bond
Controls the slow release of the peptide.
Carrier
40 kDa PEG
Increases the half-life and prevents rapid clearance.
Note: This structure is technically a prodrug because the large PEG-bound version is inactive; only the released CNP-38 peptide performs the therapeutic work.
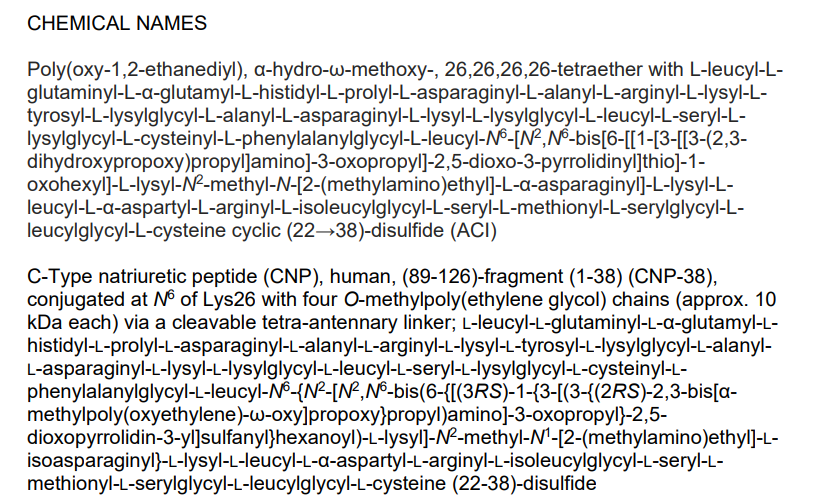
C-Type natriuretic peptide (CNP), human, (89-126)-fragment (1-38) (CNP-38), conjugated at N6 of Lys26 with four O-methylpoly(ethylene glycol) chains (approx. 10 kDa each) via a cleavable tetra-antennary linker; L-leucyl-L-glutaminyl-L-?-glutamyl-L-histid
Poly(oxy-1,2-ethanediyl), ?-hydro-?-methoxy-, 26,26,26,26-tetraether with L-leucyl-L-glutaminyl-L-?-glutamyl-L-histidyl-L-prolyl-L-asparaginyl-L-alanyl-L-arginyl-L-lysyl-L-tyrosyl-L-lysylglycyl-L-alanyl-L-asparaginyl-L-lysyl-L-lysylglycyl-L-leucyl-L-sery
FDA 2026, APPROVALS 2026, 2/27/2026, Yuviwel, Y3BH8M899D, MN-266, TRANSCON CNP, PA (224-233), Influenza, DA-66438, ACP-015, WHO 11981,
To increase linear growth in pediatric patients 2 years and older with achondroplasia with open epiphyses
Navepegritide is a prodrug consisting of a 38-amino acid C-type natriuretic peptide (CNP) moiety conjugated to a multi-arm polyethylene glycol (PEG) carrier via a cleavable linker. This structure allows for the once-weekly dosing approved by the FDA for children with achondroplasia.
Key Details
Purpose: It is designed to increase linear growth by providing continuous exposure to C-type natriuretic peptide (CNP), a protein that helps regulate bone growth.
Mechanism: As a prodrug, it uses Ascendis Pharma’s TransCon technology to release active CNP slowly into the body over a week, maintaining steady levels and avoiding high peaks.
Clinical Benefits: In the pivotal ApproaCH trial, patients treated with navepegritide showed a significant improvement in annualized growth velocity (AGV) compared to those on a placebo. It also showed potential improvements in body proportionality and lower-limb alignment.
Administration: It is administered via a once-weekly subcutaneous injection, offering a less frequent alternative to daily treatments like vosoritide.
Safety: Most common side effects include injection site reactions (redness, itching, or swelling) and a risk of low blood pressure (hypotension).
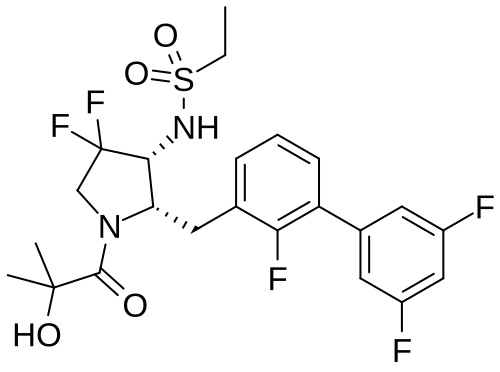
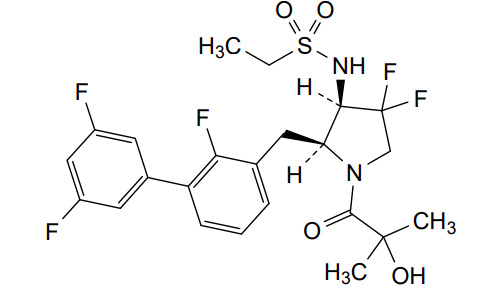
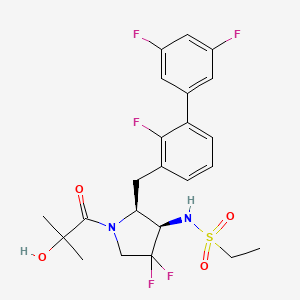
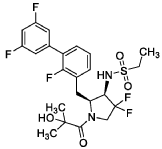
N-{(2S,3R)-4,4-difluoro-1-(2-hydroxy-2-methylpropanoyl)-2-[(2,3′,5′-trifluoro-[1,1′-biphenyl]-3-yl)methyl]pyrrolidin-3-yl}ethane-1-sulfonamide orexin type 2 receptor agonist, TAK-861, TAK 861, 59MF6P2ATF
Oveporexton is a small molecule drug. The usage of the INN stem ‘-orexton’ in the name indicates that Oveporexton is a orexin receptor agonist. Oveporexton has a monoisotopic molecular weight of 520.15 Da.
Oveporexton is being developed by Takeda.[1] As of July 2025, it has completed phase 3 clinical trials for treatment of narcolepsy, whereas no recent development has been reported for treatment of idiopathic hypersomnia.[1][5][10] Takeda plans to submit a New Drug Application (NDA) of oveporexton for the treatment of narcolepsy to the United StatesFood and Drug Administration (FDA) in 2025.[5] Oveporexton is a follow-on and replacement compound for Takeda’s earlier lead drug danavorexton (TAK-925), which is administered intravenously and stopped being developed due to unexpected liver toxicity findings.[10]
A Trial of TAK-861 for the Treatment of Narcolepsy With CataplexyCTID: NCT07363720Phase: Phase 3Status: Not yet recruitingDate: 2026-01-23Conditions: Narcolepsy Type 1 (NT1); Narcolepsy With CataplexyInterventions: PlaceboLinked Compound CID: 154617563
A Study of TAK-861 for the Treatment of Selected Central Hypersomnia ConditionsCTID: NCT05816382Phase: Phase 2/Phase 3Status: RecruitingDate: 2025-12-01Conditions: Narcolepsy Type 1Interventions: TAK-861Linked Compound CID: 154617563
A Study of TAK-861 in People With Narcolepsy Type 1CTID: NCT06505031Phase: Phase 3Status: CompletedDate: 2025-09-15Conditions: Narcolepsy Type 1Interventions: PlaceboLinked Compound CID: 154617563
A Study of TAK-861 for the Treatment of Narcolepsy Type 1CTID: NCT06470828Phase: Phase 3Status: CompletedDate: 2025-07-01Conditions: Narcolepsy Type 1Interventions: PlaceboLinked Compound CID: 154617563
A Study of TAK-861 in Participants With Narcolepsy Type 1CTID: NCT05687903Phase: Phase 2Status: CompletedDate: 2025-01-09Conditions: Narcolepsy Type 1Interventions: PlaceboLinked Compound CID: 154617563
A Randomized, Double-blind, Placebo-Controlled Study to Evaluate the Efficacy, Safety, and Tolerability of TAK-861 for the Treatment of Narcolepsy With Cataplexy (Narcolepsy Type 1)EudraCT: 2022-001654-38Phase: Phase 2Status: CompletedDate: 2023-05-26Linked Compound CID: 154617563
A Long-term Extension Study to Evaluate the Safety and Tolerability of TAK-861 in Participants With Selected Central Hypersomnia ConditionsEudraCT: 2022-002965-13Phase: Phase 2, Phase 3Status: Trial now transitionedDate: 2023-04-11Linked Compound CID: 154617563
A Randomized, Double-blind, Placebo-Controlled Study to Evaluate the Efficacy, Safety, and Tolerability of TAK-861 for the Treatment of Narcolepsy Without Cataplexy (Narcolepsy Type 2)EudraCT: 2022-002966-34Phase: Phase 2Status: CompletedDate: 2023-03-20Linked Compound CID: 154617563
A) tert-butyl (2S,3R)-3-(ethylsulfonamido)-4,4-difluoro-2-((2,3′,5′-trifluoro-[1,1′-biphenyl]-3-yl)methyl)pyrrolidine-1-carboxylate
To a mixture of tert-butyl (2S,3R)-2-(3-chloro-2-fluorobenzyl)-3-(ethylsulfonamido)-4,4-difluoropyrrolidine-1-carboxylate (3.70 g), (3,5-difluorophenyl)boronic acid (2.56 g) and 1 M aqueous potassium phosphate solution (24.3 mL) in DME (50 mL) was added XPhos Pd G3 (0.343 g) at room temperature. The mixture was stirred at 90° C. under nitrogen atmosphere for 15 h. The reaction mixture was poured into water and extracted with EtOAc. The organic layer was washed with saturated brine, dried over magnesium sulfate and concentrated under reduced pressure. The residue was purified by column chromatography (silica gel, EOAc/hexane) to give the title compound (3.30 g).
MS: [M−H] − 533.2.
B) N-((2S,3R)-4,4-difluoro-2-((2,3′,5′-trifluoro-[1,1′-biphenyl]-3-yl)methyl)pyrrolidin-3-yl)ethanesulfonamide hydrochloride
A mixture of tert-butyl (2S,3R)-3-(ethylsulfonamido)-4,4-difluoro-2-((2,3′,5′-trifluoro-[1,1′-biphenyl]-3-yl)methyl)pyrrolidine-1-carboxylate (3.30 g) and 4 M HCl/CPME solution (30 mL) was stirred overnight at room temperature. By filtration, the title compound (2.86 g) was obtained.
MS: [M+H] + 435.1.
C) N-{(2S,3R)-4,4-difluoro-1-(2-hydroxy-2-methylpropanoyl)-2-[(2,3′,5′-trifluoro[1,1′-biphenyl]-3-yl)methyl]pyrrolidin-3-yl}ethanesulfonamide
To a mixture of N-((2S,3R)-4,4-difluoro-2-((2,3′,5′-trifluoro-[1,1′-biphenyl]-3-yl)methyl)pyrrolidin-3-yl)ethanesulfonamide hydrochloride (200 mg) and DIPEA (0.367 ml) in THF (3 mL) was added alpha-acetoxy-isobutyryl chloride (0.074 ml) at 0° C., and the mixture was stirred at same temperature for 10 min. To the mixture were added water (1 ml) and 4 M lithium hydroxide solution (1.06 ml), and the mixture was stirred overnight at room temperature. The mixture was diluted with saturated brine and extracted with EtOAc. The extract was dried over magnesium sulfate and concentrated under reduced pressure. The residue was purified by column chromatography (silica gel, EtOAc/hexane) and recrystallized from EtOAc/hexane to give the title compound (154 mg).
1H NMR (400 MHz, CDCl 3) δ 1.32-1.40 (9H, m), 2.27-2.54 (1H, m), 2.88-3.16 (4H, m), 4.02-4.49 (3H, m), 4.86-5.20 (2H, m), 6.78-6.86 (1H, m), 7.02-7.10 (2H, m), 7.16-7.22 (1H, m), 7.27-7.31 (1H, m), 7.35-7.43 (1H, m).
N-{(2S,3R)-4,4-difluoro-l-(2-hydroxy-2-methylpropanoyl)-2-[(2,3′,5′-trifluoro[l,l’-biphenyl]-3-yl)methyl]pyrrolidin-3-yl}ethanesulfonamide, (hereafter referred as to “Compound A”) is described in U.S. Patent No. 11,028,048.
Dauvilliers Y, Plazzi G, Mignot E, Lammers GJ, Del Río Villegas R, Khatami R, et al. (May 2025). “Oveporexton, an Oral Orexin Receptor 2-Selective Agonist, in Narcolepsy Type 1”. The New England Journal of Medicine. 392 (19): 1905–1916. doi:10.1056/NEJMoa2405847. PMID40367374.
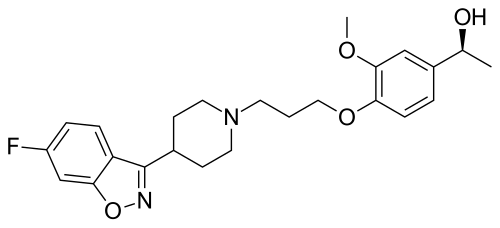
25 Feb 2026Vanda Pharmaceuticals has patent protection for an improved method of treatment with milsaperidone in USA
25 Feb 2026Vanda Pharmaceuticals has patents pending for an improved method of treatment with milsaperidone in China, Australia, Israel, Mexico and worldwide
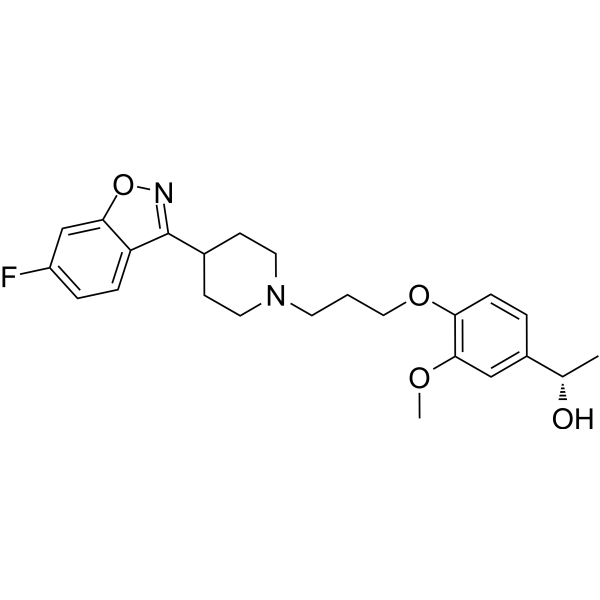
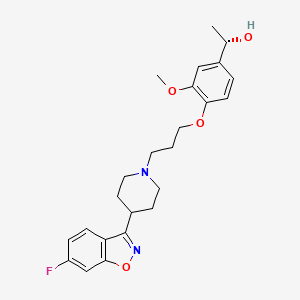
56.36 g of boran complex of (3aR, 7R)-1-methyl-3,3-diphenyl-tetrahydro-pyrrolo[1,2-c][1 ,3,2]oxazaborole (1 equivalent) is dissolved under nitrogen in methylenchloride, and the solution is cooled to 0°C. A 1M solution of 1-(4-{3-[4-(6-fluoro-benzo[d]isoxazol-3-yl)-piperidin-1-yl]-propoxy}-3-methoxy-phenyl)-ethanone (iloperidone; 1 equivalent) in methylenchloride is added via a dropping funnel over 90 minutes while the internal temperature is maintained at 0°C ± 2°C. After the addition is complete, the mixture is stirred at 0°C for 20 hours. The reaction mixture is then poured into precooled methanol (0-5°C) during 1 hour. The solution is warmed to room temperature and stirred until the H2 evolution ceases. The solution is concentrated by distillation and the residue dried in vacuum, treated with methanol and stirred for about 1 hour at 50°C and an additional hour at 0CC. The product is isolated by filtration and dried under reduced pressure for 3 hours at 50°C. The title compound is obtained (white crystals).
[α]D20– 19.3° (c=1 in chloroform) Mp: 138.2 – 138.8°C
The boran complex used as starting material can be obtained as follows:
200 ml of a solution of (3aR, 7R)-1-methyl-3,3-diphenyl-tetrahydro-pyrrolo[1,2-c][1,3,2]oxazaborole (1M in toluene) is stirred at room temperature under nitrogen. 1.2 equivalent borane-dimethylsulfide complex is added with a syringe. The solution is stirred for 2 further hours at room temperature. The borane complex is then crystallised by addition of 4 vol dry hexane and cooling to -12°C for 1.5 hour. The product is isolated by filtration in a sintered glass funnel and dried in vacuum at 40°C. The boran complex is obtained /white crystals).
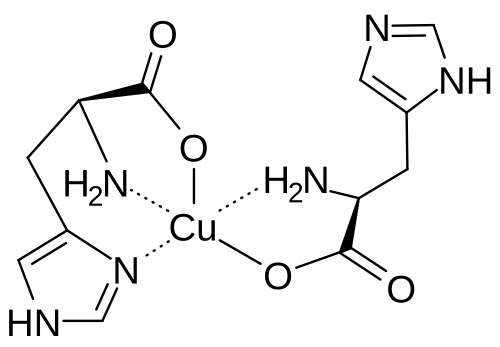
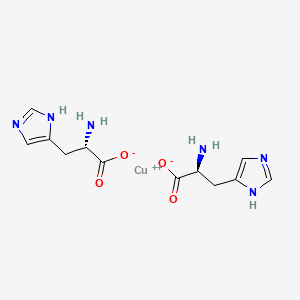
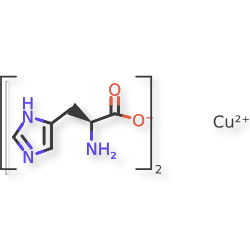
Copper histidinate, sold under the brand name Zycubo, is a medication used for the treatment of Menkes disease.[1] Copper histidinate is a copper replacement therapy given by subcutaneous injection.[1][2]
The most common side effects include infections, respiratory problems, seizures, vomiting, fever, anemia and injection site reactions.[2]
Copper histidinate was approved for medical use in the United States in January 2026.[2]
Menkes disease is a neurodegenerative disorder caused by a genetic defect that impairs a child’s ability to absorb copper.[2] The disease is characterized by seizures, failure to gain weight and grow, developmental delays, and intellectual disability.[2] It leads to abnormalities of the vascular system, bladder, bowel, bones, muscles, and nervous system.[2]
SYN
A275388 — Flores-Pulido AA, Jimenez-Perez VM, Garcia-Chong NR: Sintesis y uso de histidinato de cobre en ninos con enfermedad de Menkes en Mexico. Gac Med Mex. 2019;155(2):191-195. doi: 10.24875/GMM.18004310. [PubMed:31056589]
World Health Organization (2025). “International nonproprietary names for pharmaceutical substances (INN): recommended INN: list 94”. WHO Drug Information. 39 (3). hdl:10665/383022.
Clinical trial number NCT00811785 for “Molecular Bases of Response to Copper Treatment in Menkes Disease, Related Phenotypes, and Unexplained Copper Deficiency” at ClinicalTrials.gov
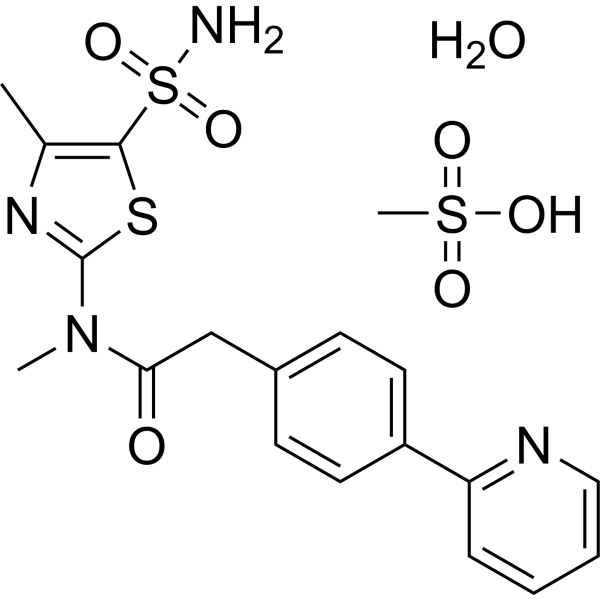
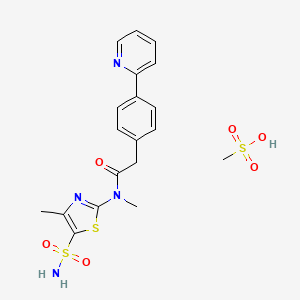
AIC316 mesylate hydrate; BAY 57-1293 mesylate hydrate
BAY57-1293; BAY 57-1293; BAY-57-1293; BAY571293; BAY 571293; BAY-571293; AIC-316; AIC 316
Hovilpri, FDA APPROVED 2026, FDA 2026
To treat adults with weakened immune systems who have mucocutaneous lesions caused by herpes simplex virus infection and have not responded to other therapies
Molecular Weight
516.61
Synonyms
AIC316 mesylate hydrate; BAY 57-1293 mesylate hydrate
Pritelivir mesylate is an antiviral drug currently under development, specifically targeting herpes simplex virus types 1 and 2 (HSV-1 and HSV-2). It functions by inhibiting the viral helicase-primase enzyme, a crucial component for HSV replication. It is being investigated as a potential treatment for various herpes infections, including those resistant to traditional antivirals like acyclovir.
Key aspects of Pritelivir mesylate:
Mechanism of Action:Pritelivir is a helicase-primase inhibitor, meaning it blocks the activity of an enzyme essential for the replication of herpes viruses.
Target Viruses:It is effective against both HSV-1 and HSV-2, the viruses responsible for cold sores and genital herpes, respectively.
Potential for Resistance:Pritelivir has shown promise in preclinical studies against acyclovir-resistant strains of HSV, making it a potential alternative for patients with drug-resistant infections.
Clinical Trials:Pritelivir is currently in phase II clinical trials, with ongoing research into its effectiveness and safety.
Route of Administration:It is being investigated for oral, topical, and vaginal administration.
Research and Development:Pritelivir is being developed by AiCuris Anti-infective Cures, building upon research from Bayer.
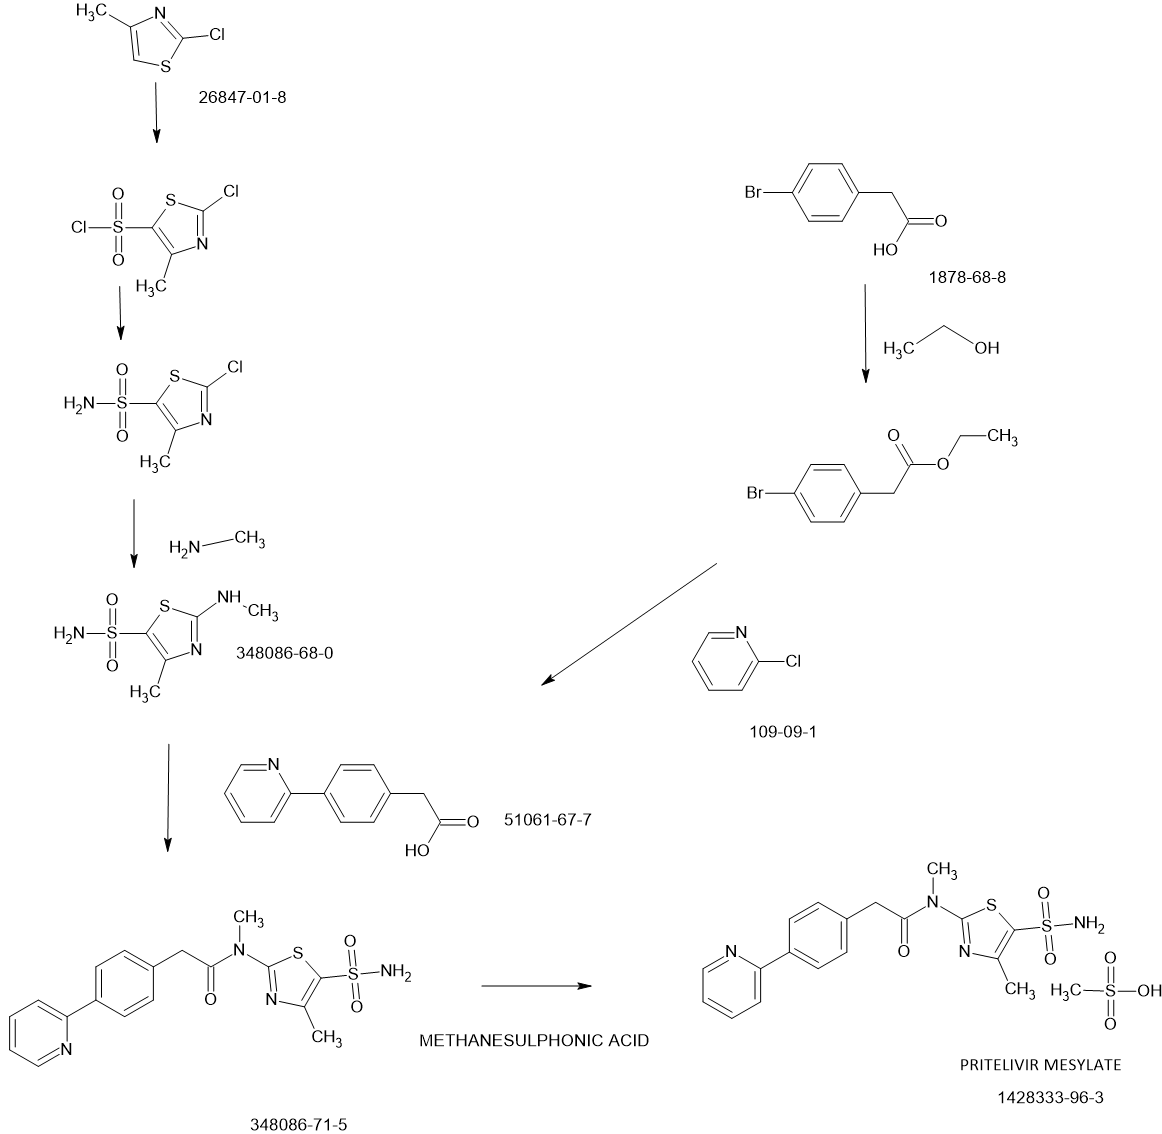
Pritelivir mesylate, an antiviral drug used to treat herpes simplex virus (HSV) infections, is synthesized through a series of chemical reactions, including palladium-catalyzed coupling, ester saponification, and amide coupling reactions. The mesylate salt is then formed by reacting the free base with methanesulfonic acid.
Detailed Synthesis Steps:
1. Diaryl Acetic Acid Synthesis:Diaryl acetic acid reagents are synthesized using palladium-catalyzed coupling reactions. These reactions involve the use of organometallic intermediates derived from halo-aryl esters.
2. Ester Saponification:The ester group in the synthesized compounds is then converted to a carboxylic acid group through saponification.
3. Amide Coupling:The resulting carboxylic acids are coupled with thiazolyl sulfonamides using amide coupling conditions to form the pritelivir molecule.
4. Salt Formation:The pritelivir free base is then reacted with methanesulfonic acid to form the mesylate salt, which is the active pharmaceutical ingredient (API).
Key Aspects of Pritelivir Mesylate Synthesis:
Targeted Mechanism:Pritelivir mesylate inhibits the herpes simplex virus by targeting the viral helicase-primase complex, essential for DNA replication, unlike traditional antivirals that target DNA polymerase.
Salt and Polymorph Screening:An extensive salt and polymorph screening is performed to optimize the pharmaceutical development of pritelivir, resulting in various salt forms including the mesylate, maleate, and sulfate.
Solubility and Stability:Pritelivir mesylate is a BCS Class II drug substance with pH-dependent solubility. It exhibits high solubility below pH 3 and poor solubility at neutral pH.
Formulation Considerations:Due to its limited water solubility, pritelivir mesylate is often formulated with solvents like DMSO, PEG300, Tween-80, and saline or with cyclodextrins like SBE-β-CD.
Clinical Trials:Pritelivir mesylate is currently under extensive study to evaluate its efficacy and safety profile, with promising results in early clinical trials.
Likewise, EP 2 598 502 Al describes the crystalline mono mesylate monohydrate salt of N- [5-(aminosulfonyl)-4-methyl- 1 ,3-thiazol-2-yl]-N-methyl-2-[4-(2-pyridinyl)phenyl]-acetar^ in a definite particle size distribution and a specific surface area range, which has demonstrated increased long term stability and release kinetics from pharmaceutical compositions, as well as to pharmaceutical compositions containing said N-[5- (aminosulfonyl)-4-methyl- 1 ,3-thiazol-2-yl]-N-methyl-2-[4-(2-pyridinyl)phenyl]acetaniide mono mesylate monohydrate having the afore-mentioned particle size distribution and specific surface area range.
WO 2013/045479 Al describes an improved and shortened synthesis process of N-[5- (ammosulfonyl)-4-methyl-l,3-thiazol-2-yl]-N-methyl-2-[4-(2-pyridinyl)phenyl]acetaniide and the mesylate salt thereof by using boronic acid derivatives or borolane reagents while avoiding toxic organic tin compounds. Moreover, also the crystalline mesylate monohydrate salt of N- [5 -(aminosulfonyl)-4-methyl- 1 ,3 -thiazol-2-yl] -N-methyl-2- [4-(2-pyridinyl)-phenyl] – acetamide is described therein with increased long-term stability and release kinetics from pharmaceutical compositions thereof.
Said pritelivir is an innovative, highly active and specific inhibitor of herpes simplex virus (HSV) infections. As a compound derived from the chemical class of thiazolylamides, pritelivir is active against both types of herpes simplex virus causing labial and genital herpes, respectively, and retains activity against viruses which have become resistant to marketed drugs. Pritelivir has a mode of action that is distinct from other antiviral agents currently in use for treatment of HSV infections (i.e., the nucleoside analogues acyclovir and its prodrug valacyclovir as well as famciclovir, the prodrug of penciclovix). Whereas nucleoside analogs terminate ongoing DNA chain elongation through inhibition of viral DNA polymerase, pritelivir prevents de novo synthesis of virus DNA through inhibition of the helicase-primase complex. In addition, it does not require activation within an HSV infected cell by viral thymidine kinase and therefore, is also protective to uninfected cells.
With the context of the invention, similar expressions which all would denote the compound pritelivir are “BAY 57-1293”, “AIC090096” and “AIC316”.
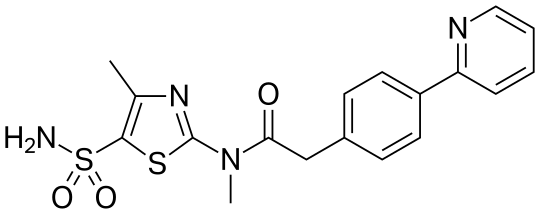
Likewise, the terms ”pritelivir”, “BAY 57-1293”, “AIC090096” and “AIC316″ or the compound *’N-[5-(ammosulfonyl)-4-methyl-l,3-thiazol-2-yl]-N-methyl-2-[4-(2-pyridm phenyl] -acetamide” would reflect throughout the text a compound having the structural formula:
Synthetic route – Manufacture of N-[5-(aminosulfonyl)-4-methyl-1,3-thiazol-2-Yl]-N-methyl- 2-[4-(2-pyridinyl)-phenyl]-acetamide free base hemihydrate
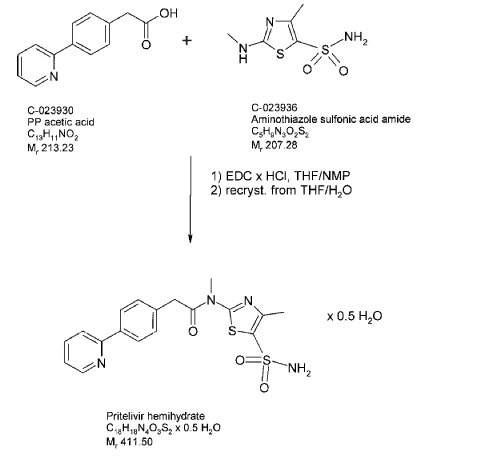
The starting materials (4-pyridine-2-yl-phenyl)-acetic acid (PP-acetic acid; C-023930) and aminothiazole sulfonic acid amide (C-023936) are coupled using standard reaction conditions
(N-Ethyl-N’-(3-dimethylaminopropyl)-carbodiimide hydrochloride (EDC x HCl), tetrahydrofuran (THF)/N-methylpyrrolidone (NMP) to deliver N-[5-(aminosulfonyl)-4- methyl-1,3-thiazol-2-yl]-N-methyl-2-[4-(2-pyridinyl)-phenyl]-acetamide free base hemihydrate (C-023931). To obtain the hemihydrate, N-[5-(aminosulfonyl)-4-methyl-1,3- thiazol-2-yl]-N-methyl-2-[4-(2-pyridinyl)-phenyl]-acetamide hemihydrate free base is recrystallized from THF/water. A flowchart showing the synthesis of N-[5-(aminosulfonyl)- 4-methyl-1,3-thiazol-2-yl]-N-methyl-2-[4-(2-pyridinyl)-phenyl]-acetamide is provided in below in the reaction scheme below 1.
Description of the manufacturing process of N-r5-(aminosulfonyl)-4-methyl-1,3-thiazol-2-yl]-N-methyl-2-[4-(2-pyridinyl)-phenyl]-acetamide free base hemihydrate
PP-acetic acid and aminothiazole sulfonic acid amide are mixed in THF/NMP, the mixture is cooled and then EDC x HCl is added in portions. The reaction mixture is stirred for several hours, and then added slowly to purified water. The suspension is stirred and filtered; the product cake is washed with purified water and dried at room temperature in a nitrogen stream and then under vacuum. Purified water is added slowly at elevated temperature, the suspension is stirred for several hours. The suspension is cooled to 5°C and stirred further for several hours. The product is isolated by filtration and washed with purified water. The product is dried at 65°C under vacuum until the criterion for water content is reached. A major advantage of the synthesis of free base of N-[5-(aminosulfonyl)-4-methyl-1,3-thiazol-2-yl]-N-methyl-2-[4-(2-pyridinyl)-phenyl]-acetamide hemihydrate is the absence impurities related to the presence of mesylate ester that might be present in the N-[5-(aminosulfonyl)-4-methyl-1,3-thiazol-2-yl]-N-methyl-2-[4-(2-pyridinyl)-phenyl]-acetamide mesylate.
PAPER
By: Carta, Fabrizio ; et al. Journal of Medicinal Chemistry (2017), 60(7), 3154-3164
SULPHURIC ACID FOR SULPHATE A SIMILAR REACTION BUT NOT SAME
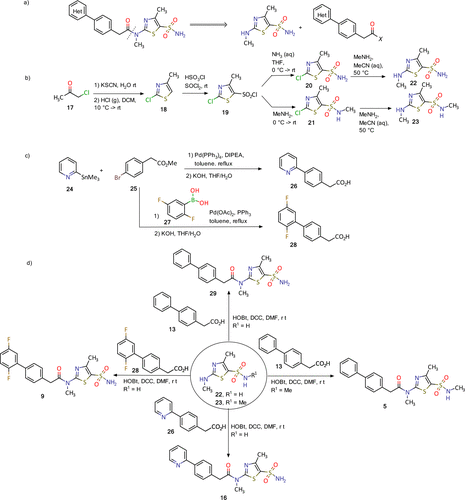
Synthesis of pritelivir and its analogues is based on the reported methods in the literature (18−20) and presented in Figure 4. A simple retrosynthetic disconnection of the target compound suggests a coupling of the thiazolyl sulfonamide and diaryl acetic acid (Figure 4a). During the course of development an optimized route applying the principles of green chemistry was developed and will be used for the commercial phase.
Figure 4. Synthesis of pritelivir (16) and analogues: (a) disconnection approach to target molecules; (b) synthesis of thiazolyl sulfonamide reagents; (c) synthesis of diaryl acetic acids; (d) synthesis of some representative examples of pritelivir and analogues. (18−20)
The synthesis of the thiazolyl sulfonamide reagents begins with a reaction of chloroacetone (17) and potassium thiocyanide to give an intermediate ketone which was cyclized to the thiazole 18 by treatment with gaseous hydrochloride (Figure 4b). Chlorosulfonylation with chlorosulfonic acid and thionyl chloride resulted in the sulfonyl chloride 19 that was converted to the corresponding sulfonamides 20 and 21 after treatment with ammonia or methylamine, respectively. The 2-chloro substituent in 20 and 21 was converted to the methyl amine in an SNAr reaction to deliver the building blocks 22–23. Diaryl acetic acid reagents were synthesized using palladium catalyzed coupling reactions with organometallic intermediates formed from the corresponding halo-aryl esters (Figure 4c). Ester saponification then delivered the corresponding carboxylic acids (e.g., 26 and 28, Figure 4c). Finally, the target molecules, for instance, 5, 11, 16, and 9, were obtained using amide coupling reaction conditions with corresponding diaryl acetic acids and the thiazolyl sulfonamides (Figure 4d). (18−20)
Medical use
Pritelivir is currently being developed for the treatment of immunocompromised patients with mucocutaneous HSV lesions that are resistant to acyclovir.
HSV in immunocompromised patients
Although HSV infection is very common in the general population, it rarely causes serious disease and is effectively contained by the immune system. In those with a weakened immune system such as transplant recipients, people receiving chemo- or radiotherapy, or HIV patients, an active HSV infection can cause disease in 35-68% of patients and may become severe or even life-threatening.[1]
Standard of care treatments
HSV treatment revolves around the use of nucleoside analogues (NA) which act via the viral DNA polymerase, causing DNA chain termination and prevention of viral replication. First-line treatment is generally acyclovir or its prodrug valacyclovir. Resistance to acyclovir is more common in HSV patients with weakened or suppressed immune systems, affecting between 4 and 25% of cases.[2][3][4][5][6]
Resistance to standard treatments
If HSV drug resistance is mediated by mutation(s) of the viral UL23 gene, which encodes the viral thymidine kinase (TK), then the pyrophosphate analogue foscarnet may be effective as a rescue treatment, since it does not require activation by TK. The use of foscarnet is commonly accompanied by restrictive toxicity, particularly nephrotoxicity.[7] If the virus also acquires resistance to foscarnet, then there is currently no FDA approved treatment.
Clinical research
Completed phase II clinical trials in otherwise healthy patients with genital herpes
A Double-blind Randomized Placebo Controlled Dose-finding Trial to Investigate Different Doses of a New Antiviral Drug in Subjects With Genital HSV Type 2 Infection.[8][9]
A Double-blind, Double Dummy, Randomized Crossover Trial to Compare the Effect of “AIC316 (Pritelivir)” 100 mg Once Daily Versus Valacyclovir 500 mg Once Daily on Genital HSV Shedding in HSV-2 Seropositive Adults.[10][11]
Ongoing phase II / phase III clinical trials with pritelivir
A phase II / III multinational, comparator-controlled, clinical trial in immunocompromised patients with acyclovir-resistant mucocutaneous lesions is listed on ClinicalTrials.gov[12] and EudraCT.[13]
Pharmacology
Mechanism of action
Pritelivir is a member of the helicase-primase inhibitors (HPI), a novel class of direct-acting antiviral drugs acting specifically against HSV-1 and HSV-2.[14][15] As the name suggests, the drugs act through inhibition of the viral helicase primase complex, encoded by the UL5 (helicase), UL8 (scaffold protein) and UL52 (primase) genes, which is essential for HSV replication.[16] The helicase primase complex is encoded separately from the viral DNA polymerase (encoded by the UL30 gene). Because HPIs i) do not target the viral DNA polymerase and ii) do not require activation by the viral thymidine kinase enzyme (encoded by the UL23 gene), mutations causing resistance to NAs are not protective against HPIs. Similarly, resistance to HPIs does not confer resistance to NAs.
^ Biswas, S.; Jennens, L.; Field, H.J. (2007). “Single amino acid substitutions in the HSV-1 helicase protein that confer resistance to the helicase-primase inhibitor BAY 57-1293 are associated with increased or decreased virus growth characteristics in tissue culture”. Arch Virol. 152 (8): 1489–500. doi:10.1007/s00705-007-0964-7. PMID17404685. S2CID23688945.
^ Field, H.J.; Biswas, S. (2011). “Antiviral drug resistance and helicase-primase inhibitors of herpes simplex virus”. Drug Resist Updat. 14 (1): 45–51. doi:10.1016/j.drup.2010.11.002. PMID21183396.
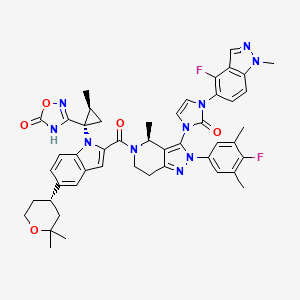
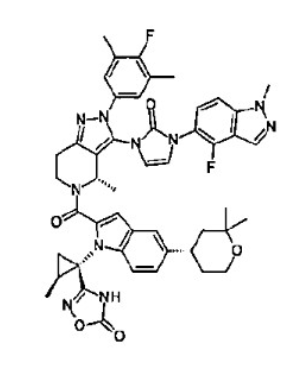
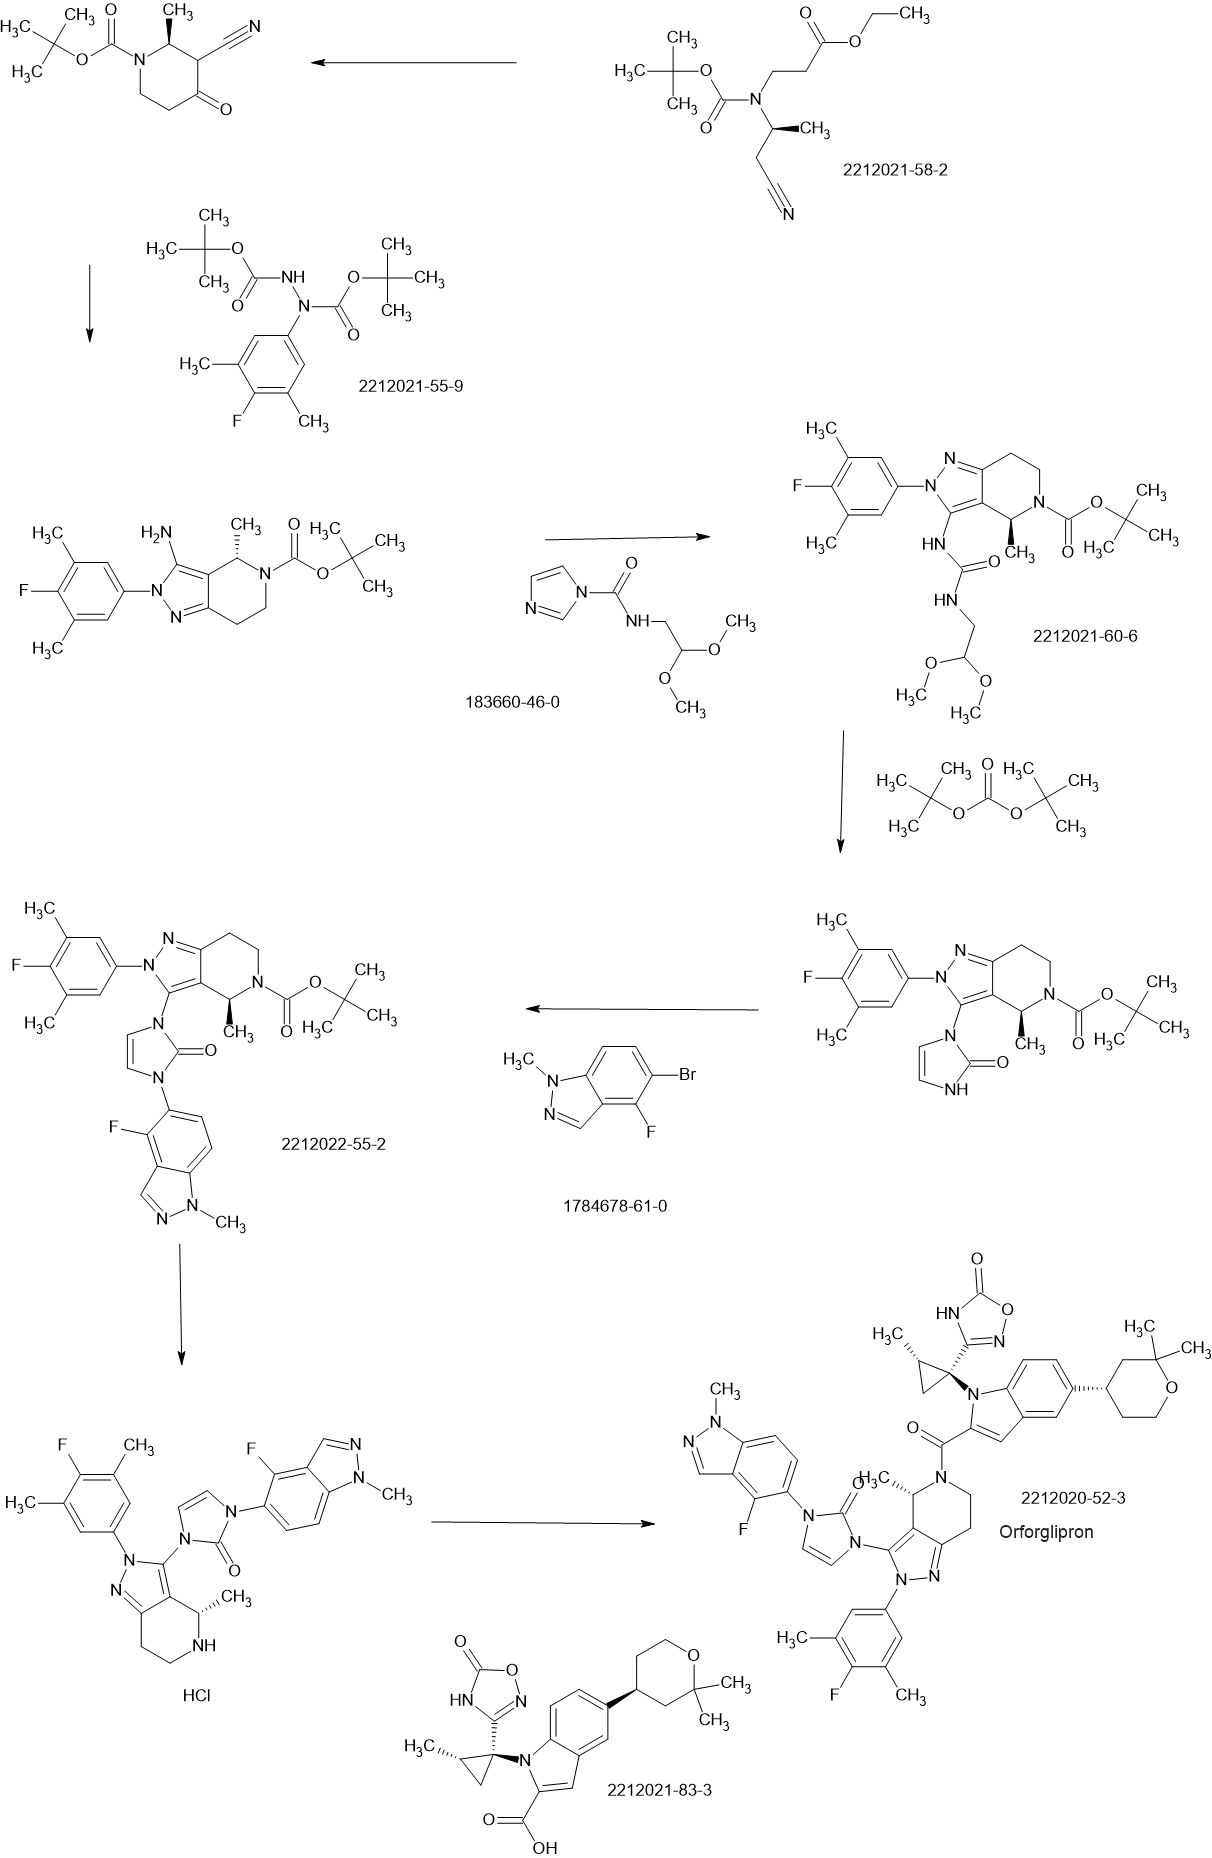
Orforglipron has a half-life of 29 to 49 hours across the doses tested and is taken once per day by mouth without food or water restrictions.[3]
Safety and dosing trials showed that the incidence of adverse events in orforglipron-treated participants was 62–89%, mostly from gastrointestinal discomfort (44–70% with orforglipron, 18% with placebo) having mild to moderate severity.[6] The most common side effects of orforglipon are diarrhea, nausea, upset stomach, and constipation.[1][6]
The ability of orforglipron to reduce blood sugar levels and body weight was judged favorable compared to dulaglutide.[6]
Phase III ACHIEVE-1 trial
In April 2025, results from a Phase III clinical trial involving 559 people with type 2 diabetes who took an oral orforglipron pill, injectabledulaglutide or a placebo daily for 40 weeks showed that orforglipron produced a reduction in blood glucose levels by 1.3 to 1.6 percentage points from a starting level of 8%.[1][7]
More than 65% of participants taking the highest dose of orforglipron achieved a reduction of hemoglobin A1C level by more than or equal to 1.5 percentage points, bringing them into the non-diabetic range as defined by the American Diabetes Association.[1] People taking the highest dose of the pill lost 8% of their weight, or around 16 lb (7.3 kg), on average after 40 weeks.[1][8]
Side effects were similar to those seen with other GLP-1 agonists, and no significant liver problems were observed.[1]
^ Wharton S, Blevins T, Connery L, et al. (June 2023). “Daily Oral GLP-1 Receptor Agonist Orforglipron for Adults with Obesity”. The New England Journal of Medicine. 389 (10): 877–888. doi:10.1056/NEJMoa2302392. PMID37351564.
To treat platinum-resistant epithelial ovarian, fallopian tube, or primary peritoneal cancer after one to three prior systemic treatment regimens, at least one of which included bevacizumab
Mechanism of ActionGlucocorticoid receptor antagonists
Orphan Drug StatusYes – Pancreatic cancer; Cushing syndrome
Phase IIICushing syndrome; Ovarian cancer; Pancreatic cancer
Phase IIFallopian tube cancer; Peritoneal cancer; Prostate cancer
Phase I/IISolid tumours
Phase IAdrenocortical carcinoma
Most Recent Events
09 Sep 2022Subgroup analysis efficacy data from a phase-II trial in Ovarian cancer presented at the 47th European Society for Medical Oncology Congress (ESMO-2022)
29 Jun 2022Phase-III clinical trials in Ovarian cancer (Combination therapy, Recurrent, Second-line therapy or greater) in USA (PO)
06 Jun 2022Corcept Therapeutics announces intentions to submit a NDA for Ovarian cancer
The drug was approved by the USFDA in 2026 for the treatment of platinum-resistant ovarian cancer.[3]
Relacorilant is an orally available antagonist of the glucocorticoid receptor (GR), with potential antineoplastic activity. Upon administration, relacorilant competitively binds to and blocks GRs. This inhibits the activity of GRs, and prevents both the translocation of the ligand-GR complexes to the nucleus and gene expression of GR-associated genes. This decreases the negative effects that result from excess levels of endogenous glucocorticoids, like those seen when tumors overproduce glucocorticoids. In addition, by binding to GRs and preventing their activity, inhibition with CORT125134 also inhibits the proliferation of GR-overexpressing cancer cells. GRs are overexpressed in certain tumor cell types and promote tumor cell proliferation.
OriginatorCorcept Therapeutics
DeveloperCorcept Therapeutics; University of Chicago
Mechanism of ActionGlucocorticoid receptor antagonists
Orphan Drug StatusYes – Pancreatic cancer; Ovarian cancer; Cushing syndrome
RegisteredFallopian tube cancer; Ovarian cancer; Peritoneal cancer
PreregistrationCushing syndrome
Phase IIIAdenocarcinoma
Phase IIProstate cancer
DiscontinuedAdrenocortical carcinoma
27 Mar 2026Discontinued – Phase-I for Adrenocortical carcinoma (Inoperable/Unresectable, Late-stage disease, Metastatic disease, Combination therapy) in USA (PO), before March 2026 (Corcept Therapeutics pipeline, March 2026)
27 Mar 2026Corcept Therapeutics plans the phase II STELLA trial for Cervical cancer (Combination therapy, Second-line therapy or greater) in first quarter of 2026
25 Mar 2026Registered for Fallopian tube cancer (Combination therapy, Second-line therapy or greater) in USA (PO) – First global approval
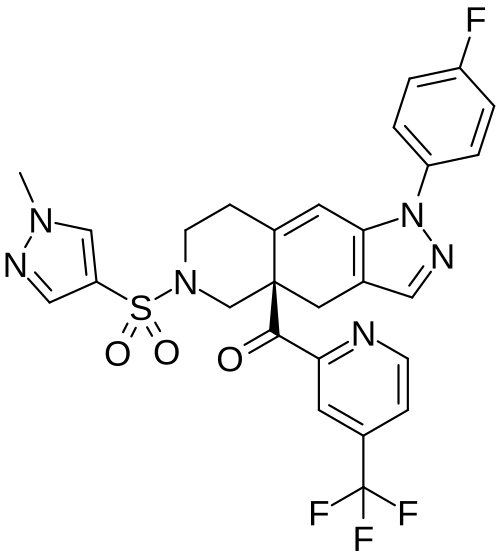
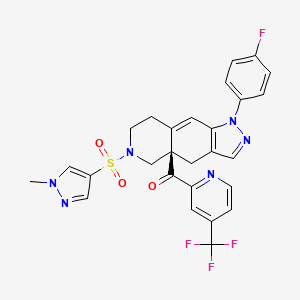
Relacorilant (CORT125134)118) is being developed by Corcept Therapeutics, Inc. It is an orally active, high-affinity, selective antagonist of the glucocorticoid receptor that may benefit from the modulation of cortisol activity. In structural optimization, the introduction of a trifluoromethyl group to the 4-position on the pyridyl moiety was found to increase HepG2 tyrosine amino transferase assay potency by a factor of four. Relacorilant is currently being evaluated in a phase II clinical study in patients with Cushing’s syndrome.119)
2-Bromo-4-(trifluoromethyl)pyridine (17) prepared from (E)-4-ethoxy-1,1,1-trifluorobut-3-en-2-one is employed as a key intermediate for the preparation of relacorilant as shown in Scheme 31.120)
118) H. Hunt, T. Johnson, N. Ray and I. Walters (Corcept Therapeutics, Inc.): PCT Int. Appl. WO2013/177559 (2013).
119) H. J. Hunt, J. K. Belanoff, I. Walters, B. Gourdet, J. Thomas, N. Barton, J. Unitt, T. Phillips, D. Swift and E. Eaton: Identification of the Clinical Candidate (R)-(1-(4-Fluorophenyl)-6-((1-methyl-1H-pyrazol-4-yl)sulfonyl)-4,4a,5,6,7,8-hexahydro-1H-pyrazolo[3,4-g]isoquinolin-4a-yl)(4-(trifluoromethyl)pyridin-2-yl)methanone (CORT125134): A Selective Glucocorticoid Receptor (GR) Antagonist. J. Med. Chem. 60, 3405–3421 (2017). [Abstract] [Google Scholar]
120) B. Lehnemann, J. Jung and A. Meudt (Archimica GmbH): PCT Int. Appl. WO 2007/000249 (2007).
The nonselective glucocorticoid receptor (GR) antagonist mifepristone has been approved in the U.S. for the treatment of selected patients with Cushing’s syndrome. While this drug is highly effective, lack of selectivity for GR leads to unwanted side effects in some patients. Optimization of the previously described fused azadecalin series of selective GR antagonists led to the identification of CORT125134, which is currently being evaluated in a phase 2 clinical study in patients with Cushing’s syndrome.
Cushing’s syndrome (CS) is a metabolic disorder caused by chronic hypercortisolism. CS is associated with cardiovascular, metabolic, skeletal and psychological dysfunctions and can be fatal if left untreated. The first-line treatment for all forms of CS is a surgery. However, medical therapy has to be chosen if surgical resection is not an option or is deemed ineffective. Currently available therapeutics are either not selective and have side effects or are only available as an injection (pasireotide).
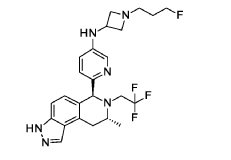
To treat hormone receptor-positive, human epidermal growth factor receptor 2-negative, locally advanced or metastatic breast cancer upon detection of ESR1 mutation during aromatase inhibitor and CDK4/6 inhibitor therapy
ClassAmines; Antineoplastics; Azetidines; Fluorinated hydrocarbons; Isoquinolines; Pyrazolones; Pyridines; Small molecules
Mechanism of ActionSelective estrogen receptor degraders
Phase IIIBreast cancer, APPROVED FDA 2026
13 Jun 2022AstraZeneca initiates a phase I drug-drug interaction trial of AZD 9833 Healthy postmenopausal female volunteers, in USA (NCT05438303)
10 Jun 2022AstraZeneca and Quotient Sciences complete the phase I QSC205863 trial in Breast cancer (In volunteers) in United Kingdom (PO, Liquid) (NCT05364255)
03 Jun 2022Safety, efficacy and pharmacokinetics data from the phase I SERENA 1 trial for Breast cancer presented at the 58th Annual Meeting of the American Society of Clinical Oncology (ASCO-2022)
Mechanism:selective estrogen receptor degrader
Area under investigation:estrogen receptor +ve breast cancer
Date commenced phase:Q1 2019
Estimated Filing Acceptance:
CountryDateUS: EU: Japan: China:
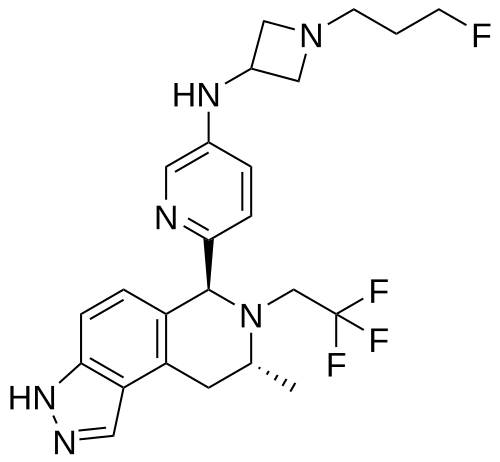
AZD9833 is an orally available selective estrogen receptor degrader (SERD), with potential antineoplastic activity. Upon administration, SERD AZD9833 binds to the estrogen receptor (ER) and induces a conformational change that results in the degradation of the receptor. This prevents ER-mediated signaling and inhibits the growth and survival of ER-expressing cancer cells
Camizestrant is an orally available selective estrogen receptor degrader (SERD), with potential antineoplastic activity. Upon administration, camizestrant binds to the estrogen receptor (ER) and induces a conformational change that results in the degradation of the receptor. This prevents ER-mediated signaling and inhibits the growth and survival of ER-expressing cancer cells
Camizestrant is an oral selective estrogen receptor degrader (ngSERD) and complete estrogen receptor (ER) antagonist.[2] It blocks the activity of estrogen receptor alpha encoded by both wild-type and mutated ESR1 and induces proteasome-dependent degradation of estrogen receptor alpha, without agonizing estrogen receptor alpha.[2]
Camizestrant was authorized for medical use in the European Union in July 2026,[2] and in the United States in September 2026.[1][5]
Discovery of AZD9833, a Potent and Orally Bioavailable Selective Estrogen Receptor Degrader and Antagonist J. Med. Chem. 2020, 63, 14530–14559, DOI: 10.1021/acs.jmedchem.0c01163.
SYN
doi: 10.1021/acs.jmedchem.0c01163.
aReagents and Conditions: (a) n-BuLi, THF, −78 oC to 0 oC, 1 h, then 4 N HCl/dioxane, RT, 1 h, 60%; (b) alkyl triflate, DIPEA, 1,4-dioxane, 90 oC, 63-74% or isobutyrylaldehyde, Na(OAc)3BH, THF, 0 oC, 56%; (c) benzophenone imine, Pd2dba3, Rac-BINAP, NaOtBu, toluene, 90 oC, then 1 N aq. HCl, 71-85%; (d) nBuLi, THF, −78 oC to 0 oC, 1 h, then 4 N HCl/dioxane, RT, 4 h; e) NH2OH, NH2OH.HCl, EtOH, reflux. 84% over 2 steps; (f) alkyl triflate, DIPEA, 1,4-dioxane, 90 oC, 44-100% or 1-fluorocyclopropane-1- carboxylic acid, HATU, Et3N, DMF, RT, 61%, then BH3.THF, THF, 65 oC, 82%.
AZD9833 is selective oestrogen receptor degrader (SERD). It works by breaking down the site where oestrogen attaches to the cancer cell. This can help stop or slow the growth of hormone receptor breast cancer. Researchers think that AZD9833 with palbociclib might work better than anastrozole and palbociclib.
AZD9833 + palbociclib
The patients will receive AZD9833 (75 mg, PO, once daily) + palbociclib (PO, once daily, 125 mg for 21 consecutive days followed by 7 days off treatment) + anastrozole placebo (1 mg, PO, once daily)
Camizestrant (AZD-9833) is a potent and orally active estrogen receptor (ER) antagonist. Camizestrant is used for the study of ER+ HER2-advanced breast cancer[1].
Camizestrant is extracted from patent US20180111931A1, example 17[1].MCE has not independently confirmed the accuracy of these methods. They are for reference only.
In Vivo
Camizestrant (oral administration; 0.2-50 mg/kg; 20 days) exhibits anti-tumour efficacy as a dose-dependent manner in human parental MCF7 mice xenograft[1]. Camizestrant (oral administration; 0.8-40 mg/kg; 30 days) decreases tumor growth as a dose-dependent manner. It gives almost complete tumour growth inhibition at the doses >10 mg/kg in mice[1]. MCE has not independently confirmed the accuracy of these methods. They are for reference only.Animal Model:Human ESR1 mutant breast cancer patient derived xenograft with CTC174 cells in female NSG mice[1]Dosage:0.8 mg/kg, 3 mg/kg, 10 mg/kg, 20 mg/kg, 40 mg/kgAdministration:Oral administration; 30 days; once dailyResult:Inhibited tumor growth in a dose-dependent manner.
Clinical Trial
NCT NumberSponsorConditionStart DatePhaseNCT04711252AstraZenecaER-Positive HER2-Negative Breast CancerJanuary 28, 2021Phase 3NCT04964934AstraZenecaER-Positive HER2-Negative Breast CancerJune 30, 2021Phase 3NCT04214288AstraZenecaAdvanced ER-Positive HER2-Negative Breast CancerApril 22, 2020Phase 2NCT04588298AstraZenecaHER2-negative Breast CancerNovember 2, 2020Phase 2NCT04541433AstraZenecaER&addition; HER2- Advanced Breast CancerSeptember 29, 2020Phase 1NCT03616587AstraZenecaER&addition; HER2- Advanced Breast CancerOctober 11, 2018Phase 1NCT04546347AstraZeneca|Quotient SciencesHealthy VolunteersSeptember 17, 2020Phase 1NCT04818632AstraZenecaER&addition;, HER2-, Metastatic Breast CancerOctober 11, 2021Phase 1
“Etcamah EPAR”. European Medicines Agency (EMA). 22 May 2026. Retrieved 30 May 2026. Text was copied from this source which is copyright European Medicines Agency. Reproduction is authorized provided the source is acknowledged.
“Etcamah PI”. Union Register of medicinal products. 22 July 2026. Retrieved 8 September 2026.
Scott JS, Moss TA, Balazs A, Barlaam B, Breed J, Carbajo RJ, et al. (December 2020). “Discovery of AZD9833, a Potent and Orally Bioavailable Selective Estrogen Receptor Degrader and Antagonist”. Journal of Medicinal Chemistry. 63 (23): 14530–14559. doi:10.1021/acs.jmedchem.0c01163. PMID32910656.
World Health Organization (2022). “International nonproprietary names for pharmaceutical substances (INN): recommended INN: list 87”. WHO Drug Information. 36 (1). hdl:10665/352794.
////////////Camizestrant, AZD 9833, AZ 14066724, UNII-JUP57A8EPZ, WHO 11592, PHASE 2, ASTRA ZENECA, CANCER, FDA 2026, APPROVALS 2026, Etcamah
Mechanism of ActionType 4 cyclic nucleotide phosphodiesterase inhibitors
RegisteredAtopic dermatitis
27 Sep 2021Registered for Atopic dermatitis (In adolescents, In children, In adults) in Japan (Topical)
11 Nov 2020Otsuka Pharmaceutical completes a phase III trial in Atopic dermatitis (In children, In adolescents, In adults) in Japan (Topical) (NCT03961529)
28 Sep 2020Preregistration for Atopic dermatitis in Japan (In children, In adolescents, In adults) (Topical)
Difamilast is under investigation in clinical trial NCT01702181 (A Safety Study to Evaluate the Use and Effectiveness of a Topical Ointment to Treat Adults With Atopic Dermatitis).
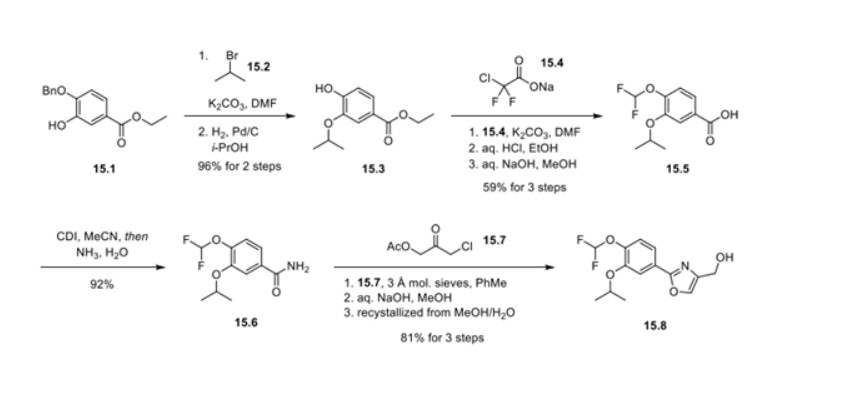
Synthesis of Oxazole Compound (Type A Crystal)
Compound (5) (white powder) was prepared in accordance with the method disclosed in Example 352 of PTL 1 (WO2007/058338).
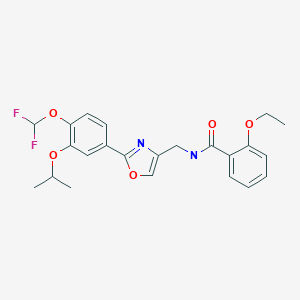
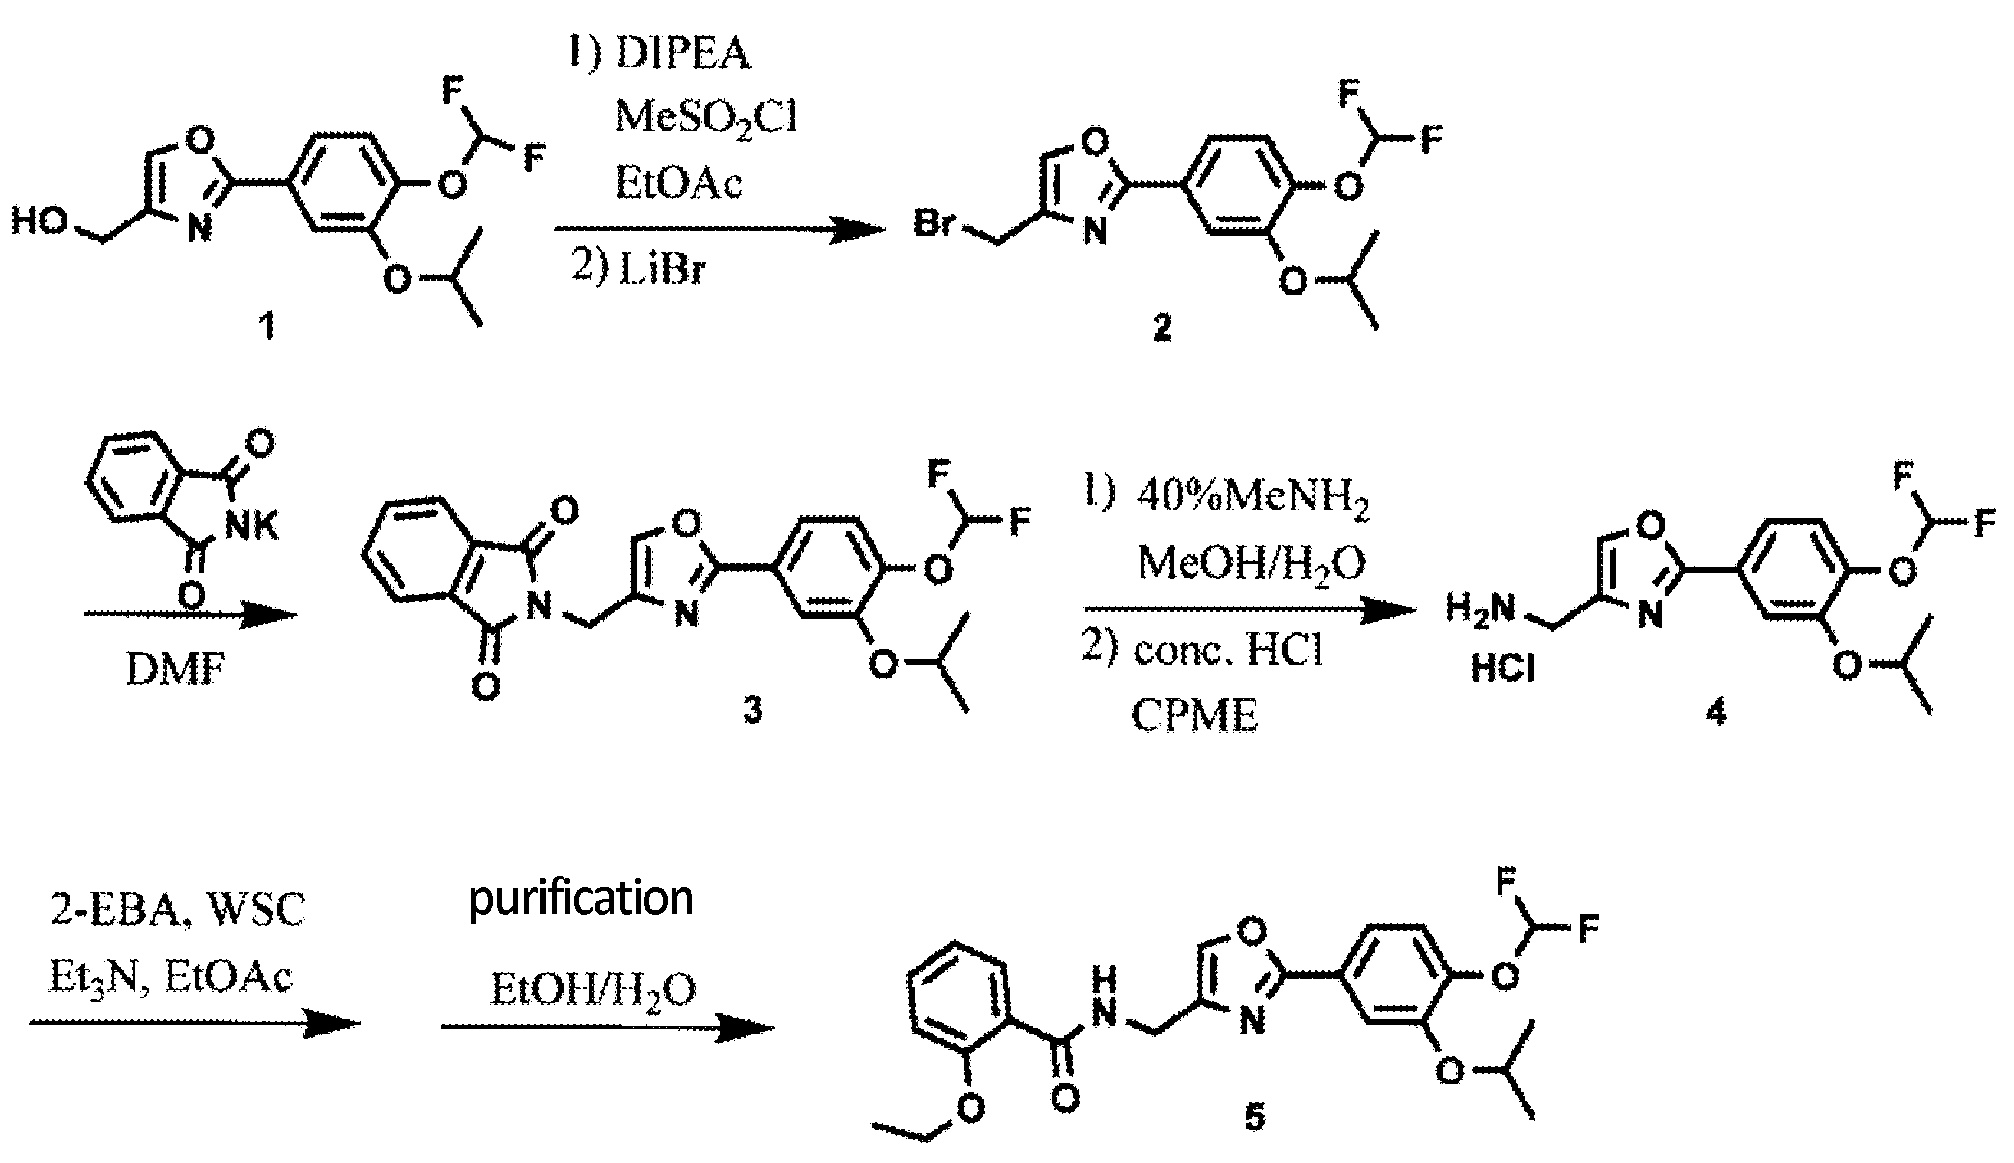
Synthesis of difamilast commenced with the monobenzylated protocatechuic acid ethyl ester 15.1. Phenol 15.1 was first converted into the corresponding isopropyl ether, which was subsequently debenzylated under palladium-catalyzed hydrogenation conditions to generate the phenolic intermediate 15.3. Difluoromethylation of 15.3 was accomplished by introducing sodium chlorodifluoroacetate 15.4 in the presence of potassium carbonate at an elevated temperature. The decarboxylative C− O bond-forming reaction presumably proceeded via a difluorocarbene species. The difluoromethylated product was treated with acid followed by ester hydrolysis under a basic medium to furnish benzoic acid derivative 15.5. Benzoic acid 15.5 was subsequently transformed into benzamide 15.6 via a benzoyl imidazole intermediate. Condensation of benzamide 15.6 with 1-acetoxy-3-chloroacetone 15.7 produced an oxazole derivative, which was subsequently saponified and recrystallized from 50% aqueous MeOH to generate alcohol 15.8.
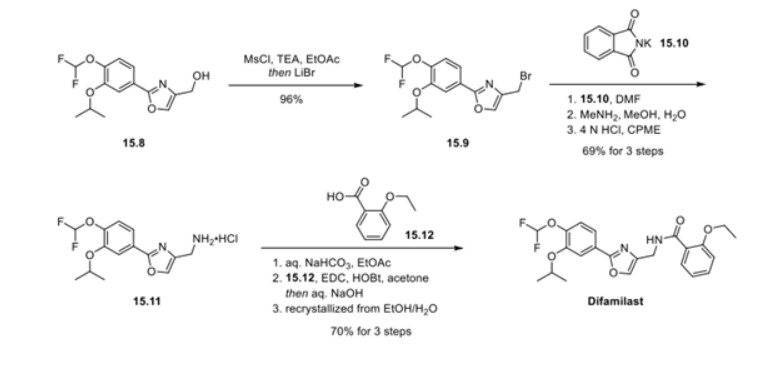
First, an activation−displacement process transformed alcohol 15.8 into bromide 15.9 via a mesylate intermediate. Alkyl bromide 15.9 was then treated with potassium phthalimide to incorporate the nitrogen center via an SN2-type displacement. Methylamine-mediated phthalimide deprotection and subsequent salt formation produced amine 15.11 as a hydrochloride salt in 69% yield over 3 steps. Finally, hydrochloride salt 15.11 was treated with aqueous sodium bicarbonate to generate a free amine, which was subjected to amide bond formation with 2-ethoxybenzoic acid 15.12 to deliver difamilast after recrystallization from aqueous EtOH.
Patent Documents 1 and 2 report an oxazole compound having a specific inhibitory action on phosphodiesterase 4 (PDE4) and a method for producing the same. PDE4 is the predominant PDE in inflammatory cells, inhibition of PDE4 increases intracellular cAMP concentration, and the increase in this concentration downregulates the inflammatory response through regulation of the expression of TNF-α, IL-23, and other inflammatory cytokines. .. Elevated cAMP levels also increase anti-inflammatory cytokines such as IL-10. Therefore, it is considered that the oxazole compound is suitable for use as an anti-inflammatory agent. For example, it may be useful for controlling skin eczema and dermatitis, including atopic dermatitis. Patent Document 3 describes an ointment that stably contains an oxazole compound having a specific inhibitory effect on PDE4 and can be efficiently absorbed into the skin. The contents of Patent Documents 1 to 3 are incorporated in the present specification by reference.
[Synthesis of Oxazole Compound (Type A Crystal)]
Compound (5) (white powder) was prepared by the method described in Example 352 of Patent Document 1 (International Publication No. 2007/088383).
[Preparation of B-type crystal 2]
Using the obtained B-type crystal as a seed crystal, it was examined to further prepare a B-type crystal. Specifically,
B-type crystals were prepared as follows according to the method described in Patent Document 3 (International Publication No. 2017/115780).
[0072]
[Chem. 6]
[0073]
Compound (1) 20.00 g (66.8 mmol) and 17.28 g (134 mmol) of diisopropylethylamine were added to 300 mL of ethyl acetate to cool the mixture, and 11.48 g (100 mmol) of methanesulfonyl chloride was introduced into the compound (1) at 10 to 30 ° C. Stir for hours. Subsequently, 17.41 g (200 mmol) of lithium bromide was added, and the mixture was stirred at 20 to 35 ° C. for 1 hour. 100 mL of water was added to the reaction solution to separate the layers, and the organic layer was concentrated under reduced pressure. 300 mL of ethyl acetate was added to the concentrated residue to dissolve it, and the mixture was concentrated again under reduced pressure. 200 mL of N, N-dimethylformamide and 17.33 g (93.6 mmol) of phthalimide potassium were added to the concentrated residue, and the mixture was reacted at 75 to 85 ° C. for 1 hour. 200 mL of water was added to the reaction solution to precipitate crystals, and the precipitated crystals were collected by filtration and dried at 80 ° C. to obtain 27.20 g (yield 95.01%) of compound (3).
[0074]
[Chem. 7]
[0075]
Compound (3) 20.00 g (46.7 mmol), 40 mL of a 40% aqueous methylamine solution, 40 mL of methanol, and 100 mL of water were mixed and reacted under reflux for 30 minutes. 200 mL of cyclopentyl methyl ether (CPME) and 20 mL of a 25% sodium hydroxide aqueous solution were added to the reaction solution, and the temperature was adjusted to 65 to 75 ° C. to separate the liquids. A mixed solution of 100 mL of water and 20.00 g of sodium chloride was added to the organic layer, and the temperature was adjusted again to 65 to 75 ° C. to separate the liquids. 5 mL of concentrated hydrochloric acid was added to the organic layer to precipitate crystals. Precipitated crystals were collected by filtration to obtain 27.58 g of wet crystals of compound (4).
[0076]
Wet crystals (46.7 mmol) of compound (4) were mixed with 120 mL of ethyl acetate and 7.1 mL (51.4 mmol) of triethylamine, and the mixture was stirred at 20 to 30 ° C. for 1 hour. To the reaction solution, 10.09 g (60.7 mmol) of 2-ethoxybenzoic acid and 11.63 g (60.7 mmol) of 1-ethyl-3- (3-dimethylaminopropyl) carbodiimide hydrochloride (WSC) were added, and 20 to 30 were added. The reaction was carried out at ° C. for 1 hour. 60 mL of water and 6 mL of concentrated hydrochloric acid were added to the reaction solution, and the temperature was adjusted to 40 to 50 ° C. to separate the solutions. 60 mL of water and 6 mL of a 25%
aqueous sodium hydroxide solution were added to the organic layer, the temperature was adjusted again to 40 to 50 ° C., the liquid was separated, and the organic layer was concentrated under reduced pressure. 50 mL of ethanol, 20 mL of water, 6 mL of a 25% aqueous sodium hydroxide solution, and 0.6 g of activated carbon were added to the concentrated residue, and the mixture was refluxed for 30 minutes. Activated carbon was removed by filtration, washed with 12 mL of ethanol, the filtrate was cooled, and 10 mg of B-type crystals (seed crystals) were added to precipitate crystals. Precipitated crystals were collected by filtration and dried at 60 ° C. to obtain 18.38 g (yield 88.18%) of crystals of compound (5).
Using the compound obtained in Example 347 and 2-bromopropane, white powdery N-[2-(4-difluoromethoxy-3-isopropoxyphenyl)oxazol-4-ylmethyl]-2-ethoxybenzamide was obtained following the procedure of Example 348.
Production Example 1: Production 1 of Compound (3)
Compound (3) was produced in accordance with the following reaction scheme.
[0146]
[Chem. 11]
[0147]
10.00 g (55.5 mmol) of compound (1a) and 9.20 g (66.6 mmol) of potassium carbonate were added to 40 ml of N,N-dimethylformamide and 6 ml of water, and the mixture was stirred until exotherm subsided. 16.92 g (111 mmol) of sodium chlorodifluoroacetate was added thereto, and the mixture was reacted at 95 to 110°C for 3 hours. 80 ml of butyl acetate and 80 ml of water were added to the reaction solution, and the solution was partitioned. 80 ml of water was added again to the organic layer, followed by partitioning. 3 ml of concentrated hydrochloric acid was added to the organic layer, and the mixture was stirred at 60 to 70°C for 30 minutes. 40 ml of water and 10 ml of a 25% sodium hydroxide aqueous solution were added to the reaction solution, and the mixture was partitioned. 5.93 g (61.1 mmol) of sulfamic acid and 10 ml of water were added to the organic layer, and 22.08 g (61.0 mmol) of a 25% sodium chlorite aqueous solution was added dropwise thereto at a temperature of 20°C or below. The mixture was reacted at 20°C or below for 15 minutes, and 10 ml of a 25% sodium hydroxide aqueous solution was added dropwise thereto at a temperature of 20°C or below, followed by pouring in 83.95 g (66.6 mmol) of a 10% sodium sulfite aqueous solution. Additionally, 2 ml of concentrated hydrochloric acid was added and the mixture was partitioned, followed by concentration of the organic layer under reduced pressure. 40 ml of methanol, 80 ml of water, and 10 ml of a 25% sodium hydroxide aqueous solution were added to the concentrated residue to dissolve the residue, and 5 ml of concentrated hydrochloric acid was added dropwise thereto to precipitate crystals. The precipitated crystals were collected by filtration and dried at 80°C, thereby obtaining 11.81 g (yield: 86.4%) of compound (3) as a white powder.
Production Example 2: Production 2 of Compound (3)
Compound (3) was produced in accordance with the following reaction scheme.
[0149]
[Chem. 12]
[0150]
10.00 g (53.2 mmol) of compound (1b), 9.55 g (69.1 mmol) of potassium carbonate, and 8.50 g (69.1 mmol) of isopropyl bromide were added to 40 ml of N,N-dimethylformamide, and the mixture was reacted at 75 to 85°C for 2 hours. 80 ml of butyl acetate and 80 ml of water were added to the reaction solution, and the mixture was partitioned. 5.68 g (58.5 mmol) of sulfamic acid and 10 ml of water were added to the organic layer, and 21.15 g (58.5 mmol) of a 25% sodium chlorite aqueous solution was added dropwise thereto at 20°C or below, followed by reaction for 15 minutes. 10 ml of a 25% sodium hydroxide aqueous solution was added thereto at 20°C or below, and subsequently 80.41 g (63.8 mmol) of a 10% sodium sulfite aqueous solution was poured in. Additionally, 2 ml of concentrated hydrochloric acid was added, and the mixture was partitioned, followed by concentration of the organic layer under reduced pressure. 40 ml of methanol, 80 ml of water, and 10 ml of a 25% sodium hydroxide aqueous solution were added to the concentrated residue, and the residue was dissolved, followed by dropwise addition of 5 ml of concentrated hydrochloric acid to precipitate crystals. The precipitated crystals were collected by filtration and dried at 80°C, thereby obtaining 12.09 g (yield: 92.4%) of compound (3) as a white powder.
[0151]
Production Example 3: Production of Compound (7)
Compound (7) was produced in accordance with the following reaction scheme.
[0152]
[Chem. 13]
Production Example 4: Production of Compound (11)
Compound (11) was produced in accordance with the following reaction scheme.
[0160]
[Chem. 14]
[0161]
Synthesis of Compound (9)
20.00 g (66.8 mmol) of compound (7) and 17.28 g (134 mmol) of N,N-diisopropylethylamine were added to 300 ml of ethyl acetate, and the mixture was cooled. 11.48 g (100 mmol) of methanesulfonyl chloride was poured in and stirred at 10 to 30°C for 1 hour. 17.41 g (200 mmol) of lithium bromide was added thereto and reacted at 20 to 35°C for 1 hour. 100 ml of water was added to the reaction solution, and the mixture was partitioned, followed by concentration of the organic layer under reduced pressure. 300 ml of ethyl acetate was added to the concentrated residue to dissolve the residue, and the solution was again concentrated under reduced pressure. 200 ml of N,N-dimethylformamide and 17.33 g (93.6 mmol) of potassium phthalimide were added to the concentrated residue and reacted at 75 to 85°C for 1 hour. 200 ml of water was added to the reaction solution to precipitate crystals. The precipitated crystals were collected by filtration and dried at 80°C, thereby obtaining 25.90 g (yield: 90.5%) of compound (9) as a white powder.
Synthesis of Compound (10)
15.00 g (35.0 mmol) of compound (9) was mixed with 30 ml of a 40% methylamine aqueous solution, 30 ml of methanol, and 75 ml of water, and reacted under reflux for 30 minutes. 150 ml of cyclopentyl methyl ether (CPME) and 15 ml of a 25% sodium hydroxide aqueous solution were added to the reaction solution, and the temperature was adjusted to 65 to 75°C, followed by partitioning. A mixture of 150 ml of water and 7.50 g of sodium chloride was added to the organic layer, and the temperature was adjusted to 65 to 75°C again, followed by partitioning. 3.75 ml of concentrated hydrochloric acid was added to the organic layer to precipitate crystals. The precipitated crystals were collected by filtration and dried at 60°C, thereby obtaining 11.95 g (yield: quant.) of compound (10) as a white powder.
Synthesis of Compound (11)
13.30 g (39.7 mmol) of compound (10) was mixed with 3.83 g (37.8 mmol) of triethylamine and 108 ml of ethyl acetate, and stirred at 20 to 30°C for 1 hour. 9.78 g (58.9 mmol) of 2-ethoxybenzoic acid and 11.28 g (58.8 mmol) of 1-ethyl-3-(3-dimethylaminopropyl)carbodiimide hydrochloride (WSC) were added to the reaction solution, and reacted at 20 to 30°C for 1 hour. 54 ml of water and 5.4 ml of concentrated hydrochloric acid were added to the reaction solution, and the temperature was adjusted to 40 to 50°C, followed by partitioning. 54 ml of water and 5.4 ml of a 25% sodium hydroxide aqueous solution were added to the organic layer, and the temperature was adjusted to 40 to 50°C again. The mixture was partitioned, and the organic layer was concentrated under reduced pressure. 45 ml of ethanol, 18 ml of water, 5.4 ml of a 25% sodium hydroxide aqueous solution, and 0.54 g of activated carbon were added to the concentrated residue, and the mixture was refluxed for 30 minutes. The activated carbon was removed by filtration, and the filtrate was washed with 11 ml of ethanol. The filtrate was cooled, and a seed crystal was added thereto to precipitate crystals. The precipitated crystals were collected by filtration and dried at 35°C, thereby obtaining 12.88 g (72.6%) of compound (11) as a white powder.
Type B Crystal Preparation 2
Analysis was conducted to further prepare the type B crystal using the obtained type B crystal as a seed crystal. More specifically, the type B crystal was prepared as follows, in accordance with the method disclosed in PTL 3 (WO2017/115780).
[0072]
[0073]
20.00 g (66.8 mmol) of compound (1) and 17.28 g (134 mmol) of diisopropylethylamine were added to 300 mL of ethyl acetate, and the mixture was cooled. 11.48 g (100 mmol) of methanesulfonyl chloride was poured in and stirred at 10 to 30°C for 1 hour. 17.41 g (200 mmol) of lithium bromide was added thereto, and the mixture was stirred at 20 to 35°C for 1 hour. 100 mL of water was added to the reaction solution, and the mixture was separated, followed by concentration of the organic layer under reduced pressure. 300 mL of ethyl acetate was added to the concentrated residue to dissolve the residue, and the solution was again concentrated under reduced pressure. 200 mL of N,N-dimethylformamide and 17.33 g (93.6 mmol) of potassium phthalimide were added to the concentrated residue, and reacted at 75 to 85°C for 1 hour. 200 mL of water was added to the reaction solution to precipitate crystals. The precipitated crystals were collected by filtration and dried at 80°C, thereby obtaining 27.20 g (yield: 95.01%) of compound (3).
[0074]
[0075]
20.00 g (46.7 mmol) of compound (3), 40 mL of a 40% methylamine aqueous solution, 40 mL of methanol, and 100 mL of water were mixed and reacted for 30 minutes under reflux. 200 mL of cyclopentyl methyl ether (CPME) and 20 mL of a 25% sodium hydroxide aqueous solution were added to the reaction solution, and the temperature was adjusted to 65 to 75°C, followed by separation. A mixture of 100 mL of water and 20.00 g of sodium chloride was added to the organic layer, and the temperature was adjusted to 65 to 75°C again, followed by separation. 5 mL of concentrated hydrochloric acid was added to the organic layer to precipitate crystals. The precipitated crystals were collected by filtration, thereby obtaining 27.58 g of compound (4) as a wet crystal.
[0076]
The wet crystal (46.7 mmol) of compound (4) was mixed with 120 mL of ethyl acetate and 7.1 mL (51.4 mmol) of triethylamine, and stirred at 20 to 30°C for 1 hour. 10.09 g (60.7 mmol) of 2-ethoxybenzoic acid and 11.63 g (60.7 mmol) of 1-ethyl-3-(3-dimethylaminopropyl)carbodiimide hydrochloride (WSC) were added to the reaction solution, and reacted at 20 to 30°C for 1 hour. 60 mL of water and 6 mL of concentrated hydrochloric acid were added to the reaction solution, and the temperature was adjusted to 40 to 50°C, followed by separation. 60 mL of water and 6 mL of a 25% sodium hydroxide aqueous solution were added to the organic layer, and the temperature was adjusted to 40 to 50°C again. The mixture was separated, and the organic layer was concentrated under reduced pressure. 50 mL of ethanol, 20 mL of water, 6 mL of a 25% sodium hydroxide aqueous solution, and 0.6 g of activated carbon were added to the concentrated residue, and the mixture was refluxed for 30 minutes. The activated carbon was removed by filtration, and the filtrate was washed with 12 mL of ethanol. The filtrate was cooled, and 10 mg of the type B crystal (a seed crystal) was added thereto to precipitate crystals. The precipitated crystals were collected by filtration and dried at 60°C, thereby obtaining 18.38 g (88.18%) of compound (5).
PATENT
WO2014034958A1
WO2007058338A2
WO2007058338A9
WO2007058338A3
US9181205B2
US2015239855A1
USRE46792E
US2020078340A1
US2017216260A1
US2019070151A1
US2009221586A1
US8637559B2
US2014100226A1
///////////
AS ON DEC2021 3,491,869 VIEWS ON BLOG WORLDREACH AVAILABLEFOR YOUR ADVERTISEMENT
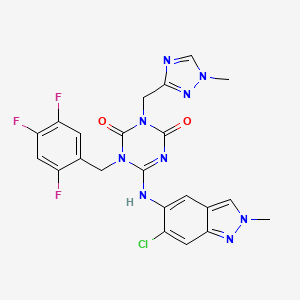
S-217622, a therapeutic drug for COVID-19, is a 3CL protease inhibitor created through joint research between Hokkaido University and Shionogi. SARS-CoV-2 has an enzyme called 3CL protease, which is essential for the replication of the virus. S-217622 suppresses the replication of SARS-CoV-2 by selectively inhibiting 3CL protease. Shionogi has already been submitting the non-clinical, manufacturing/CMC data, and clinical trial data obtained so far to the PMDA. Currently the Phase 3 part of a Phase 2/3 clinical trial in patients with mild/moderate symptoms and the Phase 2b/3 part in patients with asymptomatic/only mild symptoms are in progress.
SYN
J.Med.Chem.2024,67,4376−4418
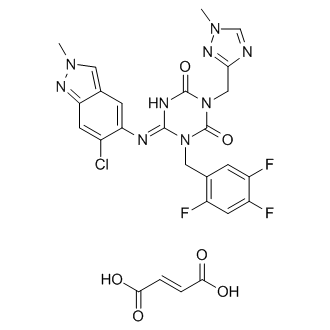
Ensitrelvir fumaric acid (3), also referred to as S-217622, is an oral noncovalent SARS-CoV-2 main protease (Mpro) inhibitordeveloped by Shionogi & Co. that was approved by the japan Pharmaceuticals and Medical Devices Agency (PMDA)for the treatment of disease caused by SARS-CoV-2 (COVID 19) infection. Dosed once daily for 5 days, ensitrelvirsuppresses the replication of SARS-CoV-2 in infected patients as a result of its inhibition of the viral mpro.25,26 Ensitrelvir retains potent inhibitory activity against many of the most common M mutants and exhibits antiviral activity against a wide variety of circulating SARS-CoV-2 variants. 27is the second Mpro Ensitrelvir inhibitor approved for the treatment of 28 disease caused by COVID-19. Unlike the first approved treatment, Paxlovid, ensitrelvir does not require coadministration with a CYP3A4 inhibitor to attenuate metabolism in vivo. Furthermore, crystal structures of ensitrelvir in complex with the main proteases of three other human-infecting coronaviruses (MERS-CoV, SARS-CoV, and HCoV-NL63)
A convergent, kilogram-scale synthesis of ensitrelvir suitable for manufacturing has been described in the literature by researchers at Shionogi. 30 The synthetic approach involved the union of two key building blocks indazole 3.7 and 1,3,5triazinone 3.14, each necessitating development of a scale worthy route. The preparation of triazinone 3.14 necessitated construction of a triazolyl methylene chloride subunit which began with the reduction of triazole ester 3.1 with aluminum hydride 3.2 (a less pyrophoric alternative to LAH yet still required aqueous Rochelle salt quench to chelate excess aluminum) 31to provide alcohol 3.3, which was then convertedto the corresponding chloride and isolated as the triazole HCl salt 3.4 (Scheme 6). Assembly of indazole intermediate 3.7began with regioselective nitration of benzaldehyde 3.5followed by treatment with hydrazine hydrate in aqueous EtOHtoprovide indazole3.6(Scheme7).Fascinatingly, the Shionogi team isolated a variety of byproducts during the conversionof3.5to3.6whichsupportedtheirhypothesisforareaction mechanism that likely equilibrated through a dibenzylidenehydrazine intermediateenroute tothedesired indazole3.6.Ascreenofelectrophilicmethyl sourcesrevealed thatMeerwein’s salt facilitatedthebest conversionof 3.6to the correspondingN2-monomethylated indazole; subsequenthydrogenative nitro reduction furnished the key indazole intermediate 3.7.Construction of the ensitrelvir core started with reaction of carboximidamide 3.8 with t-butyl isocyanate followed by N,N′carbonyldiimidazole (CDI) to secure 1,3,5-triazinone 3.10(Scheme 8). Subsequent N-alkylation with bromide 3.11provided benzyl triazinone 3.12. Substitution of the pyrazolewith m-cresol was accomplished under acidic conditions. The authors report that m-cresol was identified as a leaving group that facilitated introduction of indazole 3.7 with a minimal number of byproducts in a later step of the synthesis. The TFA-mediated reaction concomitantly removed the N-tertbutyl group providing compound 3.13 in 91% yield. Nalkylation with chloride 3.4 in the presence of a base resulted in intermediate 3.14 which was then treated with building block 3.7 in the presence of anhydrous acetic acid. Isolation of ensitrelvir fumaric acid was achieved by exposure to fumaric acid in aqueous acetone.
(25) Yotsuyanagi, H.; Ohmagari, N.; Doi, Y.; Imamura, T.; Sonoyama, T.; Ichihashi, G.; Sanaki, T.; Tsuge, Y.; Uehara, T.; Mukae, H. A phase 2/3 study of S-217622 in participants with SARS CoV-2 infection (Phase 3 part). Medicine 2023, 102, No. e33024. (26) Mukae, H.; Yotsuyanagi, H.; Ohmagari, N.; Doi, Y.; Imamura, T.; Sonoyama, T.; Fukuhara, T.; Ichihashi, G.; Sanaki, T.; Baba, K.; Takeda, Y.; Tsuge, Y.; Uehara, T. A randomized phase 2/3 study of ensitrelvir, a novel oral SARS-CoV-2 3C-like protease inhibitor, in Japanese patients with mild-to-moderate COVID-19 or asymptomatic SARS-CoV-2 infection: results of the phase 2a part. Antimicrob. Agents Chemother. 2022, 66, No. 00697. (27) Kawashima, S.; Matsui, Y.; Adachi, T.; Morikawa, Y.; Inoue, K.; Takebayashi, S.; Nobori, H.; Rokushima, M.; Tachibana, Y.; Kato, T. Ensitrelvir is effective against SARS-CoV-2 3CL protease mutants circulating globally. Biochem. Biophys. Res. Commun. 2023, 645, 132− 136. (28) Unoh, Y.; Uehara, S.; Nakahara, K.; Nobori, H.; Yamatsu, Y.; Yamamoto, S.; Maruyama, Y.; Taoda, Y.; Kasamatsu, K.; Suto, T.; et al. Discovery of S-217622, a noncovalent oral SARS-CoV-2 3CL protease inhibitor clinical candidate for treating COVID-19. J. Med. Chem. 2022, 65, 6499−6512
(29) Lin, C.; Jiang, H.; Li, W.; Zeng, P.; Zhou, X.; Zhang, J.; Li, J. Structural basis for the inhibition of coronaviral main proteases by ensitrelvir. Structure 2023, 31, 1016. (30) Kawajiri, T.; Kijima, A.; Iimuro, A.; Ohashi, E.; Yamakawa, K.; Agura, K.; Masuda, K.; Kouki, K.; Kasamatsu, K.; Yanagisawa, S.; et al. Development of a manufacturing process toward the convergent synthesis of the COVID-19 antiviral Ensitrelvir. ACS Cent. Sci. 2023, 9, 836−843. (31) Gugelchuk, M.; Silva, III, L. F.; Vasconcelos, R. S.; Quintiliano, S. A. P. Sodium bis(2-methoxyethoxy)aluminum hydride. In Encyclopedia of Reagents for Organic Synthesis; Charette, A., Bode, J., Rovis, T., Shenvi, R., Eds.; John Wiley & Sons, Ltd., 2007.
Syn
Discovery of S-217622, a Non-Covalent Oral SARS-CoV-2 3CL Protease Inhibitor Clinical Candidate for Treating COVID-19
The coronavirus disease 2019 (COVID-19) pandemic, caused by severe acute respiratory syndrome coronavirus 2 (SARS-CoV-2), has resulted in millions of deaths and threatens public health and safety. Despite the rapid global spread of COVID-19 vaccines, effective oral antiviral drugs are urgently needed. Here, we describe the discovery of S-217622, the first oral non-covalent, non-peptidic SARS-CoV-2 3CL protease inhibitor clinical candidate. S-217622 was discovered via virtual screening followed by biological screening of an in-house compound library, and optimization of the hit compound using a structure-based drug-design strategy. S-217622 exhibited antiviral activity in vitro against current outbreaking SARS-CoV-2 variants and showed favorable pharmacokinetic profiles in vivo for once-daily oral dosing. Furthermore, S-217622 dose-dependently inhibited intrapulmonary replication of SARS-CoV-2 in mice, indicating that this novel non-covalent inhibitor could be a potential oral agent for treating COVID-19.
Chemistry
The synthetic scheme for compound 1 is described in Scheme 1. Starting from the pyrazole derivative 4, cyclization with Ethyl isocyanatoacetate and CDI was conducted, giving 5 in 90% yield. Then, an alkylation with 5-bromomethyl-1,2,3-trifluorobenzene followed by introduction of a 4-difluoromethoxy-2-methylaniline unit, to give 7 (40% in 2 steps). The ester group in 7 was hydrolyzed and then amidated with methylamine, yielding 1 (58% in 2 steps). Compound 2 was synthesized similarly as shown in Scheme 2.
S-217622 (3) was synthesized as described in Scheme 3. Starting from known compound 9,21 an alkylation with 1-(bromomethyl)-2,4,5-trifluorobenzene gave 10 in 93% yield. Then, the 3-tert-Bu group was removed and the triazole unit was introduced, and the substitution of the SEt moiety with the indazole unit finally gave S-217622 (3).
21 Kai, H.; Kameyama, T.; Horiguchi, T.; Asahi, K.; Endoh, T.; Fujii, Y.; Shintani, T.; Nakamura, K.; Matsumoto, S.; Hasegawa, T.; Oohara, M.; Tada, Y.; Maki, T.; Iida, A. Preparation of triazine derivatives and pharmaceutical compound that contains same and exhibits analgesic activity. WO 2012020749 A1, Feb 16, 2012
Scheme 1.
Reagents and Conditions: (a) ethyl isocyanato-acetate, DBU, CDI, DMA, –10 °C to rt, 90%; (b) 5-bromomethyl-1,2,3-trifluorobenzene, N,N-diisopropylethylamine, DMA, 60 °C; (c) 4-difluoromethoxy-2-methylaniline, tert-butanol, 100 °C, 40% in 2 steps; (d) (i) NaOH aq., THF/MeOH, rt; (ii) methylamine, HATU, N,N-diisopropylethylamine, THF, rt., 58% in 2 steps.
Scheme 2.
Reagents and Conditions: (a) 6-chloro-2-methyl-2H-indazol-5-amine, tert-amyl alcohol, 100 °C, 44% in 2 steps from 5; (b) (i) NaOH aq., THF/MeOH, rt; (ii) methylamine, HATU, N,N-diisopropylethylamine, THF, rt., 29% in 2 steps.
Scheme 3.
Reagents and Conditions: (a) 1-(bromomethyl)-2,4,5-trifluorobenzene, K2CO3, MeCN, 80 °C, 93%; (b) TFA, rt, 97%; (c) 3-(chloromethyl)-1-methyl-1H-1,2,4-triazole hydrochloride, K2CO3, DMF, 60 °C, 45%; (d) 6-chloro-2-methyl-2H-indazol-5-amine, LHMDS, THF, 0 °C to rt., 25%.
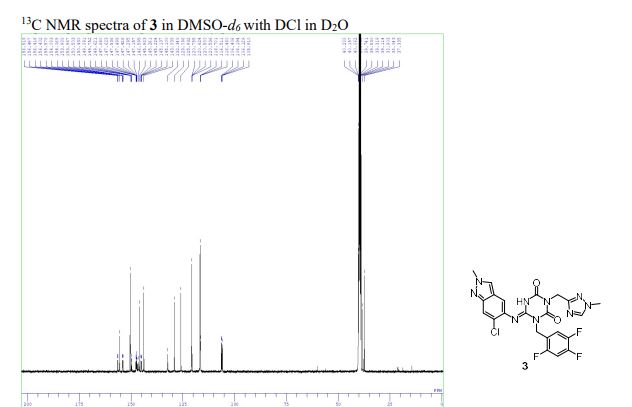
To a solution of 12 (300 mg, 0.727 mmol) and 6-chloro-2-methyl-2H-indazol-5-amine (172 mg, 0.946 mmol) in THF (6 mL) was added LHMDS (1M in THF; 1.46 mL, 1.46 mmol) dropwisely at 0 °C. The reaction mixture was stirred at 0 °C for 2.5 h and then at rt for 40 min. The reaction was quenched with aqueous NH4Cl solution, and the aqueous layer was extracted with EtOAc. The organic layer was washed with brine, dried over MgSO4, and concentrated under reduced pressure. The residue was purified by silica gel column chromatography (CHCl3/MeOH gradient, 0-20% MeOH). The solid was recrystallized from acetone/H2O to afford 3 (S-217622) (95.3 mg, 25%) as a pale brown solid. 1H NMR (400 MHz, DMSO-d6, DCl in D2O) δ 3.90 (3H, s), 4.15 (3H, s), 5.04 (2H, s), 5.26 (2H, s), 7.44 (1H, m), 7.52-7.65 (2H, m), 7.73 (1H, s), 8.40 (1H, s), 9.31 (1H, s). 13C NMR (100 MHz, DMSO-d6, DCl in D2O) δ 37.34, 38.04, 40.06, 40.29, 106.16 (dd, J = 28.2, 21.6 Hz), 116.46-116.70, 116.70, 120.54-120.76, 120.76, 125.93, 129.10, 132.35, 143.84, 145.98, 146.38 (ddd, J = 241.4, 12.5, 3.7 Hz), 146.60, 148.52 (td, J = 247.7, 13.6 Hz), 150.43, 150.50, 155.22 (ddd, J = 244.3, 10.3, 2.2 Hz), 155.58. HRMS-ESI (m/z): [M + H]+ calcd for [C22H18 F3ClN9O2]+ 532.1219; found 532.1221.
Preparation of Compound 3 (S-217622) fumaric acid co-crystal
A mixture of 3 (S-217622) (1.17 g, 2.2 mmol) and fumaric acid (278 mg, 2.4 mmol) in EtOAc (5.9 mL) was stirred at room temperature for 45 min. The suspension was filtrated to afford 3 (S-217622) fumaric acid co-crystal (1.37 g, 95 %) as a white solid. 1H NMR (400 MHz, pyridine-d5) δ 3.64 (s, 3H), 3.99 (s, 3H), 5.56 (s, 2H), 5.61 (s, 2H), 7.16-7.25 (m, 2H), 7.44 (s, 2H), 7.81 (s, 1H), 7.89 (s, 1H), 7.89-7.97 (m, 1H), 8.32 (s, 1H).
Notes
SHIONOGI has applied for a patent covering 1, 2, and 3 (S-217622). Y.U., S.U., K.N., H.N., Y.Y., S.Y., Y.M., Y.T., K.K., T.S., K.K., A.N., S.K., T.S., S.T., K.U., T.M., S.A., A.S., T.S., T.K., and Y.T. are employees of SHIONOGI & Co., Ltd. S.U., K.N., H.N., Y.M., Y.T., K.K., T.S., K.K., S.K., TS, S.T., K.U., T.S., and T.K. are shareholders in SHIONOGI & Co., Ltd. M.S., Y.O., and H.S. are financially supported by the joint research fund from SHIONOGI & Co., Ltd.
Oral antiviral medications, in addition to vaccines, are expected to play an important role in treating coronavirus disease 2019 (COVID-19), which is caused by infection with the severe acute respiratory disease coronavirus-2 (SARS-CoV-2).
These drugs must have significant antiviral activity, as well as target specificity, oral bioavailability, and metabolic stability. Although several antiviral compounds have been reported as possible SARS-CoV-2 inhibitors in vitro, only a few of these drugs have been shown to be effective in vivo.
Ensitrelvir, a novel SARS-CoV-2 antiviral
Ensitrelvir (code name S-217622, brand name Xocova), is a new inhibitor of the SARS-CoV-2 major protease (Mpro), also known as 3C-like protease, has been shown to reduce the viral load and help alleviate the severity of SARS-CoV-2 in infected hamsters. In cells, low nanomolar to sub-micromolar doses of S-217622 suppress viral growth. In hamsters, oral treatment of S-217622 showed excellent pharmacokinetic qualities and hastened recovery from acute SARS-CoV-2 infection.
S-217622 also demonstrated antiviral effectiveness against SARS-CoV-2 variants of concern (VOCs), such as the highly pathogenic Delta variant and the newly discovered Omicron variant. Overall, these findings show that S-217622, which is an antiviral drug that is currently being tested in Phase II/III clinical trials, has impressive antiviral efficiency and effectiveness against SARS-CoV-2 and could be a viable oral treatment option for COVID-19.
History
It has reached Phase III clinical trials.[3] The Japanese government is reportedly considering allowing Shionogi permission to apply for approval for medical use before the final steps of trials are completed, potentially speeding up the release for sale. This conditional early approval system has previously been used in Japan to accelerate the progression to market of other antiviral drugs targeting COVID-19, including remdesivir and molnupiravir.[6] In a study of 428 patients, viral load was reduced, but symptoms were not significantly reduced. [7]
It became the first Japanese domestic pill to treat COVID-19, third to be regulatorally approved in Japan; in February 2022.[8]
 DRUG APPROVALS BY DR ANTHONY MELVIN CRASTO
.....FOR BLOG HOME CLICK HERE


 Subscribe in a reader






 This article incorporates text from this source, which is in the public domain.