New Drug Approvals

Home » Posts tagged 'Antineoplastic' (Page 13)

Tag Archives: Antineoplastic

DRUG APPROVALS BY DR ANTHONY MELVIN CRASTO .....FOR BLOG HOME CLICK HERE

Blog Stats

  • 5,087,729 hits

Flag and hits

Flag Counter

Enter your email address to follow this blog and receive notifications of new posts by email.

Join 37.7K other subscribers
Follow New Drug Approvals on WordPress.com

Archives

Categories

Recent Posts

Flag Counter

ORGANIC SPECTROSCOPY

Read all about Organic Spectroscopy on ORGANIC SPECTROSCOPY INTERNATIONAL 

Enter your email address to follow this blog and receive notifications of new posts by email.

Join 37.7K other subscribers
DR ANTHONY MELVIN CRASTO Ph.D

DR ANTHONY MELVIN CRASTO Ph.D

DR ANTHONY MELVIN CRASTO, Born in Mumbai in 1964 and graduated from Mumbai University, Completed his Ph.D from ICT, 1991,Matunga, Mumbai, India, in Organic Chemistry, The thesis topic was Synthesis of Novel Pyrethroid Analogues, Currently he is working with AFRICURE PHARMA, ROW2TECH, NIPER-G, Department of Pharmaceuticals, Ministry of Chemicals and Fertilizers, Govt. of India as ADVISOR, earlier assignment was with GLENMARK LIFE SCIENCES LTD, as CONSUlTANT, Retired from GLENMARK in Jan2022 Research Centre as Principal Scientist, Process Research (bulk actives) at Mahape, Navi Mumbai, India. Total Industry exp 32 plus yrs, Prior to joining Glenmark, he has worked with major multinationals like Hoechst Marion Roussel, now Sanofi, Searle India Ltd, now RPG lifesciences, etc. He has worked with notable scientists like Dr K Nagarajan, Dr Ralph Stapel, Prof S Seshadri, etc, He did custom synthesis for major multinationals in his career like BASF, Novartis, Sanofi, etc., He has worked in Discovery, Natural products, Bulk drugs, Generics, Intermediates, Fine chemicals, Neutraceuticals, GMP, Scaleups, etc, he is now helping millions, has 9 million plus hits on Google on all Organic chemistry websites. His friends call him Open superstar worlddrugtracker. His New Drug Approvals, Green Chemistry International, All about drugs, Eurekamoments, Organic spectroscopy international, etc in organic chemistry are some most read blogs He has hands on experience in initiation and developing novel routes for drug molecules and implementation them on commercial scale over a 32 PLUS year tenure till date Feb 2023, Around 35 plus products in his career. He has good knowledge of IPM, GMP, Regulatory aspects, he has several International patents published worldwide . He has good proficiency in Technology transfer, Spectroscopy, Stereochemistry, Synthesis, Polymorphism etc., He suffered a paralytic stroke/ Acute Transverse mylitis in Dec 2007 and is 90 %Paralysed, He is bound to a wheelchair, this seems to have injected feul in him to help chemists all around the world, he is more active than before and is pushing boundaries, He has 100 million plus hits on Google, 2.5 lakh plus connections on all networking sites, 100 Lakh plus views on dozen plus blogs, 227 countries, 7 continents, He makes himself available to all, contact him on +91 9323115463, email amcrasto@gmail.com, Twitter, @amcrasto , He lives and will die for his family, 90% paralysis cannot kill his soul., Notably he has 38 lakh plus views on New Drug Approvals Blog in 227 countries......https://newdrugapprovals.wordpress.com/ , He appreciates the help he gets from one and all, Friends, Family, Glenmark, Readers, Wellwishers, Doctors, Drug authorities, His Contacts, Physiotherapist, etc He has total of 32 International and Indian awards

Verified Services

View Full Profile →

Archives

Categories

Flag Counter

IMATINIB


Imatinib3Dan.gif

Imatinib2DACS.svg

Imatinib

CAS No:- [152459-95-5]

IUPAC Name:- 4-[(4-Methyl-1-piperazinyl)methyl]-N-[4-methyl-3-[[4-(3-pyridinyl)-2-pyrimidinyl]amino]phenyl]benzamide

M. P.:- 211-213 °C

MW: 493.604

4-[(4-methylpiperazin-1-yl)methyl]-N-(4-methyl-3-{[4-(pyridin-3-yl)pyrimidin-2-yl]amino}phenyl)benzamide

-[(4-methylpiperazin-1-yl)methyl]-N-(4-methyl-3-{[4-(pyridin-3-yl)pyrimidin-2-yl]amino}phenyl)benzamide

N-(4-methyl-3-((4-(pyridin-3-yl)pyrimidin-2-yl)amino)phenyl)-4-((4-methylpiperazin-1-
yl)methyl)benzamide


Imatinib (INN), marketed by Novartis as Gleevec (Canada, South Africa and the USA) or Glivec (Australia, Europe and Latin America), and sometimes referred to by its investigational name STI-571, is a tyrosine-kinase inhibitor used in the treatment of multiple cancers, most notably Philadelphia chromosome-positive (Ph+) chronic myelogenous leukemia (CML).[1]

Like all tyrosine-kinase inhibitors, imatinib works by preventing a tyrosine kinase enzyme, in this case BCR-Abl, fromphosphorylating subsequent proteins and initiating the signalling cascade necessary for cancer growth and survival, thus preventing the growth of cancer cells and leading to their death by apoptosis.[2] Because the BCR-Abl tyrosine kinase enzyme exists only in cancer cells and not in healthy cells, imatinib works as a form of targeted therapy—only cancer cells are killed through the drug’s action.[3] In this regard, imatinib was one of the first cancer therapies to show the potential for such targeted action, and is often cited as a paradigm for research in cancer therapeutics.[4]

Imatinib has been cited as the first of the exceptionally expensive cancer drugs, costing $92,000 a year. Doctors and patients complain that this is excessive, given that its development costs have been recovered many times over, and that the costs of synthesizing the drug are orders of magnitude lower. In the USA, the patent protecting the active principle will expire on 4 January 2015 while the patent protecting the beta crystal form of the active principal ingredient will expire on 23 May 2019.[5]

The developers of imatinib were awarded the Lasker Award in 2009 and the Japan Prize in 2012.[6][7]

bcr-abl kinase (green), which causes CML, inhibited by imatinib (red; small molecule).

Medical uses

Imatinib is used to treat chronic myelogenous leukemia (CML), gastrointestinal stromal tumors (GISTs) and a number of othermalignancies.

Chronic myelogenous leukemia

The U.S. Food and Drug Administration (FDA) has approved imatinib as first-line treatment for Philadelphia chromosome-positive CML, both in adults and children. The drug is approved in multiple Philadelphia chromosome-positive cases of CML, including after stem cell transplant, in blast crisis, and newly diagnosed.[8]

Gastrointestinal stromal tumors

The FDA first granted approval for advanced GIST patients in 2002. On 1 February 2012, imatinib was approved for use after the surgical removal of KIT-positive tumors to help prevent recurrence.[9] The drug is also approved in unresectable KIT-positive GISTs.[8]

Other

The FDA has approved imatinib for use in adult patients with relapsed or refractory Philadelphia chromosome-positive acute lymphoblastic leukemia (Ph+ ALL), myelodysplastic/ myeloproliferative diseases associated with platelet-derived growth factor receptor gene rearrangements, aggressive systemic mastocytosis without or an unknown D816V c-KIT mutation, hypereosinophilic syndrome and/or chronic eosinophilic leukemia who have the FIP1L1-PDGFRα fusion kinase (CHIC2 allele deletion) or FIP1L1-PDGFRα fusion kinase negative or unknown, unresectable, recurrent and/or metastaticdermatofibrosarcoma protuberans.[8] On 25 January 2013, Gleevec was approved for use in children with Ph+ ALL.[10]

For treatment of progressive plexiform neurofibromas associated with neurofibromatosis type I, early research has shown potential for using the c-KIT tyrosine kinase blocking properties of imatinib.[11][12][13][14]

Legal challenge to generics

In 2007, imatinib became a test case through which Novartis challenged India’s patent laws. A win for Novartis would make it harder for Indian companies to produce generic versions of drugs still manufactured under patent elsewhere in the world. Doctors Without Borders argues a change in law would make it impossible for Indian companies to produce cheap generic antiretrovirals (anti-HIV medication), thus making it impossible for Third World countries to buy these essential medicines.[43] On 6 August 2007, the Madras High Court dismissed the writ petition filed by Novartis challenging the constitutionality of Section 3(d) of Indian Patent Act, and deferred to the World Trade Organization (WTO) forum to resolve the TRIPS compliance question. As of 2009 India has refused to grant patent exclusivity..

On April 01, 2013 Supreme Court of India dismissed the plea of Novartis for the grant of patent.

in germany

Mechanism of action

Mechanism of action of imatinib
Imatinib
Drug mechanism
1IEP.png

Crystallographic structure of tyrosine-protein kinase ABL (rainbow colored, N-terminus = blue, C-terminus = red) complexed with imatinib (spheres, carbon = white, oxygen = red, nitrogen = blue).[31]
Therapeutic use chronic myelogenous leukemia
Biological target ABL, c-kit, PDGF-R
Mechanism of action Tyrosine-kinase inhibitor
External links
ATC code L01XE01
PDB ligand id STI: PDBe, RCSB PDB
LIGPLOT 1iep

Imatinib is a 2-phenyl amino pyrimidine derivative that functions as a specific inhibitor of a number of tyrosine kinase enzymes. It occupies the TK active site, leading to a decrease in activity.

There are a large number of TK enzymes in the body, including the insulin receptor. Imatinib is specific for the TK domain inabl(the Abelson proto-oncogene), c-kit and PDGF-R (platelet-derived growth factorreceptor).

In chronic myelogenous leukemia, the Philadelphia chromosome leads to a fusion protein of abl with bcr(breakpoint cluster region), termed bcr-abl. As this is now aconstitutively active tyrosine kinase, imatinib is used to decrease bcr-abl activity.

The active sites of tyrosine kinases each have a binding site for ATP. The enzymatic activity catalyzed by a tyrosine kinase is the transfer of the terminal phosphate from ATP to tyrosine residues on its substrates, a process known as protein tyrosinephosphorylation. Imatinib works by binding close to the ATP binding site of bcr-abl, locking it in a closed or self-inhibited conformation, and therefore inhibiting the enzyme activity of the protein semi-competitively.[32] This fact explains why many BCR-ABL mutations can cause resistance to imatinib by shifting its equilibrium toward the open or active conformation.[33]

Imatinib is quite selective for bcr-abl – it does also inhibit other targets mentioned above (c-kit and PDGF-R), but no other knowntyrosine kinases. Imatinib also inhibits the abl protein of non-cancer cells but cells normally have additional redundant tyrosine kinases which allow them to continue to function even if abl tyrosine kinase is inhibited. Some tumor cells, however, have a dependence on bcr-abl.[34] Inhibition of the bcr-abl tyrosine kinase also stimulates its entry in to the nucleus, where it is unable to perform any of its normal anti-apoptopic functions.[35]

The Bcr-Abl pathway has many downstream pathways including the Ras/MapK pathway, which leads to increased proliferation due to increased growth factor-independent cell growth. It also affects the Src/Pax/Fak/Rac pathway. This affects the cytoskeleton, which leads to increased cell motility and decreased adhesion. The PI/PI3K/AKT/BCL-2 pathway is also affected. BCL-2 is responsible for keeping the mitochondria stable; this suppresses cell death by apoptosis and increases survival. The last pathway that Bcr-Abl affects is the JAK/STAT pathway, which is responsible for proliferation.[36]

synthesis

…………………………

Imatinib is known as an inhibitor of protein-tyrosine kinase and is indicated for the treatment of chronic myeloid leukemia (CML). Imatinib also has potential for the treatment of various other cancers that express these kinase including acute lymphocyte leukemia and certain solid tumors. It can also be used for the treatment of atherosclerosis, thrombosis, restenosis, or fibrosis. Thus, imatinib can also be used for the treatment of non-malignant diseases. Imatinib is usually administered orally in the form of a suitable salt, e.g., in the form of imatinib mesylate.

The chemical name of Imatinib is 4-(4-methyl piperazine -1- methyl) -N-4-methyl-3-[4- (3- pyridyl) pyrimidine-2-amino] – benzamide and is represented by the following structural formula:

Figure imgf000003_0001

(Imatinib)

Imatinib Mesylate is an inhibitor of signal transduction (STI571) invented by Novartis AG after 7 years of hard work; it is the first inhibitor of cancer signal transduction ratified in the whole world. It is sold by Novartis as Gleevec capsules containing imatinib mesylate in amounts equivalent to 100 mg or 400 mg of imatinib free base.

Imatinib Mesylate is the rare drug in America, European Union and Japan. In May 10, 2001, it was ratified by American Food and Drug Administration (FDA) to treat the chronic myelogenous leukemia patients. EP0564409 (US5521 184) describes the process for the preparation of imatinib and the use thereof, especially as an anti tumour agent.

There are generally two synthetic routes for synthesis of Imatinib, suitable for the industrial production. One synthetic process as described in scheme-I comprises using 2-methyl-5-nitroaniline as the raw material which is reacted with cyanamide to obtain guanidine; cyclization reaction with 3-dimethylamino-l-(3-pyridyl)-2-propylene-l- ketone; reduction step of nitro to amine and condensation reaction with 4- (Chloromethyl)benzoyl chloride and N-methylpiperazidine to obtain Imatinib (WO 2004/108669). -I

Figure imgf000004_0001

Scheme-2 describes the successful process for the synthesis of Imatinib using 4-methyl-3- nitroanilines as the raw material, comprising reacting 4-methyl-3-nitroanilines with 4- (Chloromethyl)benzoyl chloride and N-methyl piperazidine in turns; followed by reduction of nitro group to amino group; then reaction with cyanamide to obtain guanidine; finally cyclization reaction with 3- dimethyl amino- 1 -(3- pyridyl)-2- propylene-1 -ketone to obtain Imatinib (WO 03/066613). The said PCT application discloses the use of 4-4-(methyl piperazin-l-ylmethyl)-benzoic acid methyl ester as one of the raw material but rest of the reactants are different from that of N-(5-amino -2- methylphenyl)-4-(3-pyridyl)-2-pyrimidine amine in presence of trimethyl aluminium.

Scheme-2

Figure imgf000005_0001

Common feature of the processes for preparing imatinib according to (WO 2004/108669) and (WO03/066613) lies in use of cyanamide as a reagent. The main difference between the two routes is that the reaction sequence of cyclization of pyrimidine chain is different. Example 10 of PCT International Publication no. WO 2003/066613 is less applicable to industrial purposes. These include the reaction between N-(3-bromo-4-methyl-phenyl)-4- (4-methyl-piperazin-l -ylmethyl)-benzamide and 4-(3-pyridyl)-2-pyrimidineamine which uses a mixture of rac-BINAP (a phosphine oxide catalyst) and Pd2 (dba)3*CHCl3. These catalysts are very expensive, therefore, their use is unfit for commercial production.

CN1630648A describes a process comprising reaction of 3- bromine-4- methyl aniline with 4-(4-methyl-piperazin- methyl) methyl benzoate in presence of trimethyl-Aluminum to obtain N-(4-methyl-3-bromobenzene)-4-(4-methyl-piperazin- 1 -methyl)-benzamide, which further reacts with 2-amino-4-(3-pyridyl)- pyrimidine in presence of palladium as catalyst to obtain Imatinib.

Figure imgf000006_0001
Figure imgf000006_0002

The drawback of the above process is the use of trimethyl-Aluminum, which is flammable and reacts severely when comes in contact with water.

CN101016293A describes another process using N-(4-methyl-3-3- aminophenyl)-4-(4- methyl-piperazin-1 -methyl)- benzamide as the raw material. The said raw material is reacted with 2-halogen-4-(3-pyridyl)- pyrimidine to obtain Imatinib.

Figure imgf000006_0003

The process disclosed in CN 101016293 A comprises use of halogenated agent, such as phosphorus oxychloride, which is used to synthesize 2-halogeno-4- methyl- (3-pyridyl) – pyridine is lachrymator and corrosive and has great influence to the surroundings. EP0564409 describes a coupling reaction between N-(5-amino -2-methylphenyl)-4-(3- pyridyl)-2-pyrimidine amine and 4-(4-methyl piperazin-l-ylmethyl)-benzoyl chloride in the presence of high quantity of pyridine to starting reactant amine N-(5-amino -2- methylphenyl)-4-(3-pyridyl)-2-pyrimidine amine. The ratio of the pyridine to the said reactant is 138 which is equivalent to about 40 parts v/w. Use of such a large quantity of pyridine is unsafe as it is a toxic solvent according to ICH guidelines. The workup of the reaction comprises evaporation of the remaining pyridine and further processing, which finally involves chromatography for purification, which is highly undesirable on industrial scale because it is expensive and time consuming.

Figure imgf000007_0001

US2006/0149061 and US20060223817 also discloses a similar synthetic approach comprising the use of similar pyridine /starting amine ratio (140 equivalents which is equals about 41 parts v/w). The product obtained is purified by slurring in ethyl acetate.

WO2004/074502 describes a coupling reaction between N-(5-amino -2-methylphenyl)-4- (3-pyridyl)-2-pyrimidine amine and 4-(4-methyl piperazin-l-ylmethyl)-benzoyl chloride wherein solvent like dimethyl pharmamide , dimethyl acetamide, N-methyl pyrilidinone are used as solvents instead of pyridine. However the method described in this patent application lacks an advantage in that the coupling reaction produces the hydrohalide salt of imatinib, e.g. imatinib trihydrochloride monohydrate, which has to be treated with a base in order to afford the imatinib base, thus an extra step is required. Further, conventional methods for coupling N-(5-amino -2-methylphenyl)-4-(3-pyridyl)-2- pyrimidine amine require reaction with an acid halide, e.g. 4-(4-methyl piperazin-1- ylmethyl)-benzoyl chloride, which requires an additional production step that can involve harsh and/or toxic chlorinating agent.

Figure imgf000008_0001

WO2008/1 17298 describes a coupling reaction between N-(5-amino -2-methylphenyl)-4- (3-pyridyl)-2-pyrimidine amine and 4-(4-methyl piperazin-l-ylmethyl)-benzoyl chloride in presence of a base selected from potassium carbonate, sodium carbonate, potassium or sodium hydroxide. Use of potassium carbonate as base results into the formation of Imatinib dihydrochloride which ultimately requires an additional operation of neutralization by using excessive base to get imatinib.

Figure imgf000008_0002

WO2008/136010 describes a coupling reaction between N-(5-amino -2-methylphenyl)-4- (3-pyridyl)-2-pyrimidine amine and 4-(4-methyl piperazin-l-ylmethyl)-benzoyl chloride in presence of base potassium hydroxide resulting into 78.6% yield of crude imatinib base. Preparation of crude requires imatinib hydrochloride preparation during the workup which is then basified to get base in crude form. This also describes maleate salt preparation as mode of purification which is again basified to give pure Imatinib base.

Figure imgf000009_0001

US patent application 2004/0248918 discloses a different approach using N-(5-amino -2- methylphenyl)-4-(3-pyridyl)-2-pyrimidine amine and 4-(2-chloromethyl)-benzoyl chloride as raw material. The reaction for the preparation of Imatinib is carried out in tetrahydrofuran as a reaction solvent and in the presence of pyridine as a base. However the method described in this patent application lacks an advantage as purification of the product requires column chromatography using chloroform: methanol (3: 1 v/v), which is not suitable purification method when performing the reaction on large scale, followed by crystallizati

Figure imgf000009_0002

US patent application 2008/0103305 discloses a process comprising reacting N-(5-amino -2-methylphenyl)-4-(3-pyridyl)-2-pyrimidine amine or its alkyl derivative and an acid salt of 4-[(4-methyl-l-piperazinyl)-methyl] benzoyl derivative as given below in the scheme-3 using pyridine in an amount of about 2 to 10 volumes per gram of the said amine. However the drawback associated with this process is use of pyridine especially when reaction is performed on large scale. -3

Figure imgf000010_0001

………………………….

SYNTHESIS

Anticancer drug imatinib mesylate (Imatinib <wbr> Mesylate)

Inverse synthetic analysis will be divided into four imatinib into fragment A has 1,3 – parents electrical, fragment B are 1,3 – parent nuclear, fragments A and B constitute a pyrimidine ring.

Anticancer drug imatinib mesylate (Imatinib <wbr> Mesylate)

Compound 4 can be obtained in two ways, benzyl bromide 1 and secondary amines 2 by SN2 reaction, or the aldehyde 3 with a secondary amine 2 by reductive amination. Sodium cyanoborohydride electron withdrawing effect of the cyano group, thereby reducing the activity of the negative hydrogen, it may be present in acidic solution. Also in the acidic conditions of aldehydes and secondary amines imine positive ions, which is higher than the activity of aldehyde reduction.This is why the reductive amination reagent with inert negative and hydrogen under acidic conditions. 4 hydrolyzed ester with thionyl chloride into the acid chloride 5 . The reaction of aniline and cyanamide dinucleophile guanidine 7 . Compound 8 and DMF-DMA reaction electrophilic reagent parents 9 , 7 , and 9 ring closure under alkaline conditions to generate 10 . Finally, reduction, amidation, and a salt of imatinib mesylate generated.

………………………………..

Org. Process Res. Dev., 2012, 16 (11), pp 1794–1804
DOI: 10.1021/op300212u
Abstract Image

An efficient, economic process has been developed for the production of imatinib with 99.99% purity and 50% overall yield from four steps. Formation and control of all possible impurities is described. The synthesis comprises the condensation of N-(5-amino-2-methylphenyl)-4-(3-pyridinyl)-2-pyrimidineamine with 4-(4-methylpiperazinomethyl)benzoyl chloride in isopropyl alcohol solvent in the presence of potassium carbonate to yield imatinib base.

…………………………

Org. Biomol. Chem., 2013,11, 1766-1800

DOI: 10.1039/C2OB27003J

http://pubs.rsc.org/en/content/articlelanding/2013/ob/c2ob27003j#!divAbstract

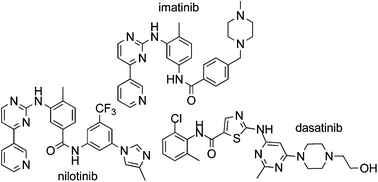
Imatinib (1), nilotinib (2) and dasatinib (3) are Bcr-Abl tyrosine kinase inhibitors approved for the treatment of chronic myelogenous leukemia (CML). This review collates information from the journal and patent literature to provide a comprehensive reference source of the different synthetic methods used to prepare the aforementioned active pharmaceutical ingredients (API’s).

Graphical abstract: The synthesis of Bcr-Abl inhibiting anticancer pharmaceutical agents imatinib, nilotinib and dasatinib

……………………..

Organic Process Research & Development, 12(3), 490-495. DOI: 10.1021/op700270nAs an example of research aimed at industrial production one involving imatinib. This cancer drug was one of the first offspring of rational drug design and if you believe the Wikipedia page hugely expensive despite its simple appearance (no stereocenters!). A group of Northwest University researchers set out to improve the existing Novartis procedure DOI and here is how they did it.

2-acetylpyridine (1) was alkylated with the acetal of N,N-dimethylformamide 2 to enamine 3. A pyrimidine ring in 5was formed with base and reagent guanidine nitrate 4 and nitrotoluene fragment 6 was added in a Ullmann-type reaction with CuI generating secondary amine 7. The nitro group was reduced by hydrazine / FeCl3/C to the amine which was then converted to amide 8 with acid chloride 9. The final step is addition of piperazine 10 to form imatinib11.So is this procedure an improvement on the existing method and ready-made for industrial implementation? Surely they have eradicated the use of toxic cyanamide, cumbersome sodium metal and expensive palladium but they have also introduced equally toxic hydrazine and the harmful and explosive guanidine nitrate. As a further point of criticism the final step is demonstrated on a 0.5 gram scale. If the journal Organic Process Research & Development would live up to its standards the scale would at least be a kilogram.Liu, Y., Wang, C., Bai, Y., Han, N., Jiao, J., Qi, X. (2008). A Facile Total Synthesis of Imatinib Base and Its Analogues. Organic Process Research & Development, 12(3), 490-495. DOI: 10.1021/op700270n
………………………………………….

Tetrahedron Lett. 2007, 48, 3455. DOI: 10.1016/j.tetlet.2007.03.033

Angelo Carotti and his group from University of Bari have developed a solid-phase synthesis of Imatinib which acts as a selective tyrosine kinase inhibitor (Tetrahedron Lett. 2007, 48, 3455. DOI: 10.1016/j.tetlet.2007.03.033). By applying microwave heating in five steps of the synthesis (preparation of linker 1, nucleophilic substitution, reduction of the nitro group, formation of guanidine and final cyclization) the total process could be accelerated. Key steps were the guanylation of aniline 2 where a higher yield and purity of product 3 could be obtained under microwave irradiation, and the final cyclization to resin bound Imatinib where the reaction time could be reduced from 20 h to 50 min. Addiotionally, resin stability was ensured due to the shorter reaction time.

………………………………

…………………………………….

http://www.google.com/patents/EP2509973A1?cl=en

process for the preparation of imatinib, which comprises the reaction of 4-Methyl-N-(4-pyridin-3-yl-pyrimidin-2-yl)- benzene-l,3-diamine (II) also referred as N-(5-amino -2-methylphenyl)-4-(3-pyridyl)-2- pyrimidine amine with 4-(4-Methyl-piperazin-l-ylmethyl)-benzoic acid ester (III) in the presence of a base in a suitable solvent to yield substantially pure imatinib base in about 90% yield.

Figure imgf000012_0001

R is C1-C4 alkyl group The preparation of 4-Methyl-N-(4-pyridin-3-yl-pyrimidin-2-yl)-benzene-l,3-diamine (II) and 4-(4-Methyl-piperazin-l-ylmethyl)-benzoic acid ester (III) may be carried out according to prior art methods.

Compound of formula (II) can be synthesized by the process disclosed in WO2004/ 108669 comprising

Figure imgf000012_0002

reacting 2-methyl-5-nitroaniline with 50% aqueous solution of cyanamide to obtain N-(2- Methyl-5-nitrophenyl)-guanidinium nitrate, which further reacted with 3-dimethylamino- l-pyridin-3-yI-propenone to yield (2-methyI-5-nitrophenyl)-(4-pyridin-3-yI-pyrimidin -2- yl)-amine, finally, reduction of nitro group to obtain compound of formula (Π).

Componds of formula (III) can be synthesized by the process disclosed in synthtic communications 2003, 3597

Figure imgf000013_0001

comprising reacting a-halogen-/?-toluinitrile or methanesulfonic acid 4-cyano-benzyl ester or toluene-4-sulfonic acid 4-cyano-benzyl ester with N-methylpiperazine, followed by hydrolysis of the cyano to acid which formed as dihydrochloride contain half crystalline hydrate, finally reaction with alcohol to obtain compound of formula (III).

The synthetic route for preparing imatinib according to the present invention is is given below

Figure imgf000013_0002

EXAMPLES

Example 1

To a solution of 4-Methyl-N-(4-pyridin-3-yl-pyrimidin-2-yl)-benzene-l,3-diamine (27.7g) and 4-(4-Methyl-piperazin-l-ylmethyl)-benzoic acid methyl ester (50g) in Tetrahydrofuran (250ml), a solution of sodium methylate (lOg) in methanol (10ml) was added. The reaction mixture was heated to reflux. After completion of the reaction solution was poured into ice-water and a large amount of solid precipitated, which was filtered and washed with water and dried to obtain Imatinib base (45g). Yield: 91%.

The spectral data is as follows:

Ή NMR ( 500M , DMSO ) δ : 10.2 (s, lH), 9.30 (s, 1H), 8.99 (s, 1H), 8.72 (d, J=4.0

Hz, 1H), 8.57 (s, 1H), 8.53 (s, 1H), 8.11 (s, 1H), 8.00 (s, 1H), 7.98 (s, 1H), 7.58-7.51 (m, 4H), 7.44 (d, J=4.3 Hz, 1H), 7.22 (d, J=8.1 Hz, 1H), 3.70 (s, 2H), 3.50-3.25 (m, 2H),

3.20-2.90 (m, 4H), 2.81 (s, 3H), 2.40 (s, 3H), 2.24 (s, 3H). 13C NMR (125M ,

DMSO ) δ : 164.9, 161.3, 161.1, 159.4, 150.8, 147.7, 137.7, 137.1, 134.9, 134.3, 132.3, 129.9, 129.1, 127.7, 127.6, 123.9, 117.2, 1 16.8, 107.5, 59.9, 52.1, 48.9, 42.2, 17.5.

MS (M++l): 494.3

Example 2

To a solution of 4-Methyl-N-(4-pyridin-3-yl-pyrimidin-2-yl)-benzene-l,3-diamine (27.7g) and 4-(4-Methyl-piperazin-l-ylmethyl)-benzoic acid methyl ester (50g) in toluene (250ml), a solution of sodium ethoxide (20g) in methanol (10ml) was added. The reaction mixture was heated to reflux. After completion of the reaction, solution was poured into ice-water and a large amount of solid precipitated, which was filtered and washed with water and dried to obtain Imatinib base (44g). Yield: 91%.

Example 3

To a solution of potassium butoxide (250g) in methanol (1000ml), a solution of 4-Methyl- N-(4-pyridin-3-yl-pyrimidin-2-yl)-benzene-l,3-diamine (277g) and 4-(4-Methyl- piperazin-l-ylmethyl)-benzoic acid propyl ester (600g) in Tetrahydrofuran (2500ml) was added. The reaction mixture was stirred at room temperature. After completion of the reaction solution was poured into ice-water and a large amount of solid precipitated, which was filtered and washed with water and dried to obtain Imatinib base (450g). Yield: 91%. Example 4

To a solution of potassium butoxide (25kg) in ethanol (lOOLitre), a solution of 4-Methyl- N-(4-pyridin-3-yI-pyrimidin-2-yl)-benzene-l,3-diamine (27.7kg) and 4-(4-MethyI- piperazin-l-ylmethyl)-benzoic acid ethyl ester (50.0kg) in toluene (250Litre) was added. The reaction mixture was stirred at room temperature. After completion of reaction, solution was poured into ice-water and a large amount of solid precipitated, which was filtered and washed with water, and dried to obtain Imatinib base (40.0kg). Yield: 81%.

…………………………………

FIGURE 2.

FIGURE 2.

Synthesis of SKI696. (A) isopropanol/sodium hydroxide (a); iron/acetic acid/EtOH/water (b); triethyl amine/acetonitrile (c); 1-ethyl-3-(3-dimethylaminopropyl)carbodiimide hydrochloride/N,N-dimethylaminopyridine/dimethylformamide (d); trifluoroacetic acid/dichloromethane (e); potassium carbonate/acetonitrile (f). (B) 18F-KF/Kryptofix/1,2-dichlorobenzene (g); dimethylformamide/acetonitrile (h).

http://jnm.snmjournals.org/content/52/8/1301/F2.expansion.html

………………………………….

http://www.beilstein-journals.org/bjoc/single/articleFullText.htm?publicId=1860-5397-9-265

In order to prepare the core heterocyclic unit a direct condensation between a 1,3-dicarbonyl compound 3.39 and an amidine or guanidine 3.40 is frequently employed (Scheme 36a). Alternatively, an amidine can be condensed with a vinylogous amide 3.41 resulting in the direct formation of 2,4-disubstituted pyrimidines. These condensations often require relatively harsh reaction conditions despite this they are of great value as they involve cheap or easily accessible materials and typically only form water as the principle byproduct.

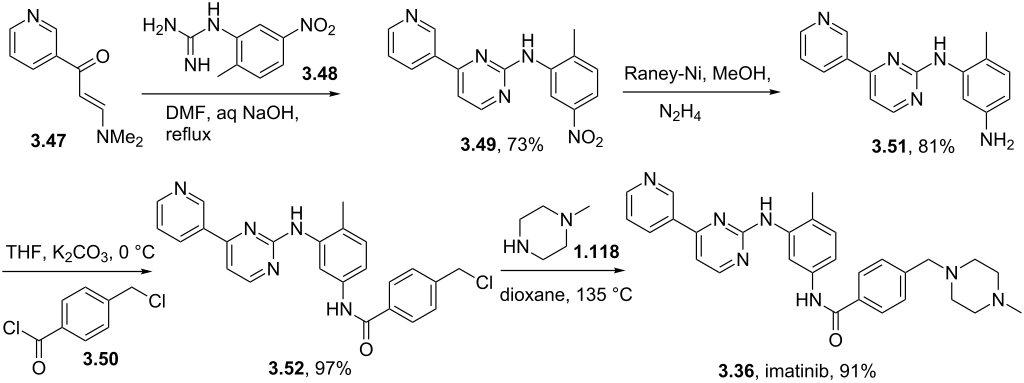
A modification of the above pyrimidine synthesis has been applied in the generation of imatinib (3.36, Gleevec) which is Novartis’ tyrosine kinase inhibitor used for the treatment of chronic myeloic leukaemia. In a patented route the aldol product 3.47 undergoes a condensation reaction with guanidine 3.48 in basic media to give the 2-aminopyrimidine 3.49 (Scheme 37) [93]. After generating the functional pyrimidine core a hydrazine-mediated reduction of the nitro group in the side chain was conducted with Raney-Nickel as the catalyst. Amide formation with 4-chloromethylbenzoyl chloride (3.50) and a direct displacement of the benzylic chloride with N-methylpiperazine (1.118) complete this synthesis of imatinib in excellent overall yields.

[1860-5397-9-265-i37]
Scheme 37: Synthesis of imatinib.

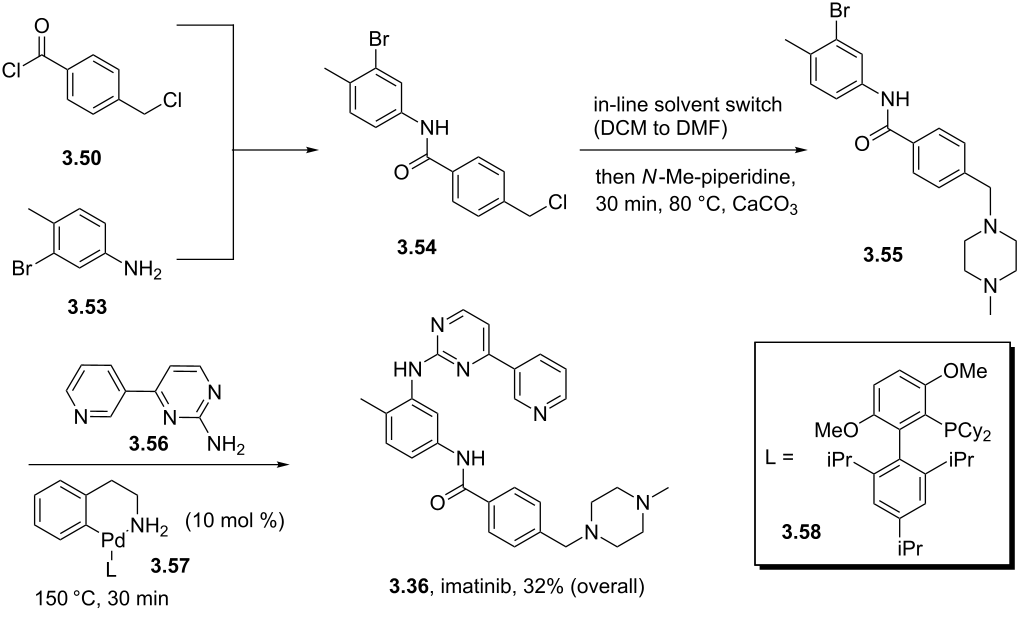
One noteworthy feature of this imatinib synthesis is that it is specifically designed for facile isolation of intermediates by precipitation due to their limited solubility in non-polar solvents [94]. Whilst this process was efficient in enabling the isolation of pure material after each step, it does not encourage telescoping of steps, which would in principal increase the overall efficiency of the process. Recently, similar approaches have been utilised in the academic environment using enabling techniques in a route to imatinib. For instance, our group has employed continuous flow synthesis methods to imatinib [95,96]. The route not only afforded imatinib but led to many previously inaccessible derivatives in an automated fashion within a single working day (Scheme 38). In addition, this particular sequence showcases the uses of scavenger resins for in-line purification as the synthesis progresses and features the use of a Buchwald–Hartwig amination in a late stage fragment coupling. While it was sufficient to access only small amounts of these structures (around 50 mg), these techniques are currently being adopted by several major pharmaceutical companies in order to enhance drug development and even manufacturing sequences.

[1860-5397-9-265-i38]
Scheme 38: Flow synthesis of imatinib.
pick up ref from
http://www.beilstein-journals.org/bjoc/single/articleFullText.htm?publicId=1860-5397-9-265

…………………………………..

Imatinib is a tyrosine-kinase inhibitor used for the treatment of cancer.  A key steps in its synthesis is the enamine formation highlighted in green below.

1) Show a mechanism for this transformation?

2) This particular enamine is rather stable.  Comment on it relatively high stability?  (i.e. What make it so stable?)

Imatinib Synthesis

……………………………………..

Org. Biomol. Chem., 2009,7, 5129-5136

DOI: 10.1039/B913333J

http://pubs.rsc.org/en/content/articlelanding/2009/ob/b913333j#!divAbstract

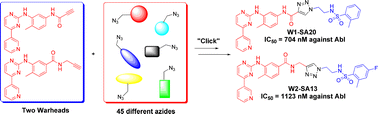
Protein kinases catalyze the phosphorylation of serine, threonine, tyrosine and histidine residues in proteins. Aberrant regulation of kinase activity has been implicated in many diseases including cancer. Thus development of new strategies for kinase inhibitor design remains an active area of research with direct relevance to drug development. Abelson (Abl)tyrosine kinase is one of the Src-family of tyrosine kinases and is directly implicated in Chronic Myelogenous Leukemia (CML). In this article, we have, for the first time, developed an efficient method for the construction of small molecule-based bisubstrate inhibitors of Abl kinase using click chemistry. Subsequent biochemical screenings revealed a set of moderately potentinhibitors, a few of which have comparable potency to Imatinib (an FDA-approved drug for treatment of chronic myeloid leukemia) against Abl.

Graphical abstract: Rapid synthesis of Abelson tyrosine kinase inhibitors using click chemistry

……………………………………

Medicine for Blood Cancer

‘Imitinef Mercilet’ is a medicine which cures blood cancer.
Its available free of cost at “Adyar Cancer Institute in Chennai”.
Create Awareness. It might help someone.Cancer Institute in Adyar, Chennai

‘Imitinef Mercilet’ is apparently an alternative spelling of the drug Imatinib mesylate which is used in the treatment of some forms of leukemia along with other types of cancer. Imatinib, often referred to a “Gleevec”, has proved to be an effective treatment for some forms of cancers. However, “blood cancer” is a generalized term for cancers that affect the blood, lymphatic system or bone marrow. The three types of blood cancer are listed as leukemia, lymphoma, and multiple myeloma. These three malignancies require quite different kinds of treatments. While drugs (including Imatinib), along with other treatments such as radiation can help to slow or even stop the progress of these cancers, there is currently no single drug treatment that can be said to actually cure all such cancers.

Category: Cancer
Address: East Canal Bank Road , Gandhi Nagar
Adyar, Chennai -600020
Landmark: Near Michael School
Phone: 044-24910754 044-24910754 ,
044-24911526 044-24911526 , 044-22350241

Imatinib is a small molecule selectively inhibiting specific tyrosine kinases that has emerged recently as a valuable treatment for patients with advanced GIST. The use of imatinib as monotherapy for the treatment of GIST has been described in PCT publication WO 02/34727, which is here incorporated by reference. However, it has been reported that primary resistance to imatinib is present in a population of patients, for example 13.7% of patients in one study. In addition, a number of patients acquire resistance to treatment with imatinib. More generally this resistance is partial with progression in some lesions, but continuing disease control in other lesions. Hence, these patients remain on imatinib treatment but with a clear need for additional or alternative therapy.

Imatinib is 4-(4-methylpiperazin-1-ylmethyl)-N-[4-methyl-3-(4-pyridin-3-yl)pyrimidin-2-ylamino)phenyl]-benzamide having the formula I

The preparation of imatinib and the use thereof, especially as an anti-tumour agent, are described in Example 21 of European patent application EP-A-0 564 409, which was published on 6 Oct. 1993, and in equivalent applications and patents in numerous other countries, e.g. in U.S. Pat. No. 5,521,184 and in Japanese patent 2706682

flow synthesis

The flow-based route required minimal manual intervention and was achieved despite poor solubility of many reaction components

21 January 2013Michael Parkin

UK chemists have used a combination of flow chemistry methods with solid-supported scavengers and reagents to synthesise the active pharmaceutical ingredient, imatinib, of the anticancer drug Gleevec. The method avoids the need for any manual handling of intermediates and allows the drug to be synthesised in high purity in less than a day.

Gleevec, developed by Novartis, is a tyrosine kinase inhibitor used for the treatment of chronic myeloid leukaemia and gastrointestinal stromal tumours.

READ ALL AT

http://www.rsc.org/chemistryworld/2013/01/flow-synthesis-anticancer-drug

IMATINIB

CREDIT

http://www.veomed.com/va041542042010

‘Wrapping’ Gleevec Fights Drug-Resistant Cancer, Study Shows

 http://www.sciencedaily.com/releases/2007/05/070501115127.htm

The anti-cancer drug Gleevec® is far more effective against a drug-resistant strain of cancer when the drug wraps the target with a molecular bandage that seals out water from a critical area. This image shows the bandage (black box) on the modified version of the drug, WBZ-7. (Credit: Image courtesy of Rice University)

A new study in Cancer Research finds that the anti-cancer drug Gleevec® is far more effective against a drug-resistant strain of cancer when the drug wraps the target with a molecular bandage that seals out water from a critical area.

FIG 23.8 Optimization of imatinib as a chemotherapeutic agent. The discovery that 2-phenylaminopyrimidine inhibitors of PKC also inhibit the unrelated v-Abl oncogene turned attention to its potential use in the treatment of chronic myelogenous leukaemia. Starting with the 2-phenylaminopyrimidine backbone, addition of the benzamidine group increased activity against tyrosine kinases, the methyl group reduced its activity against PKC (so-called ‘ target hopping ’ ). Addition of a 3’-pyridyl group improved the activity in cellular assays. Subsequent addition of N -methylpiperazine increased water solubility and oral bioavailability, enabling the drug to survive the stomach and to enter the bloodstream.

 ……………………..

An automated flow-based synthesis of imatinib: the API of gleevec M.D. Hopkin, I.R. Baxendale, S.V. Ley, J.C.S. Chem. Commun.2010, 46, 2450-2452.

References

  1. Jump up^ Novartis Pharma AG. Gleevec® (imatinib mesylate) tablets prescribing information. East Hanover, NJ; 2006 Sep. Anon. Drugs of choice for cancer. Treat Guidel Med Lett. 2003; 1:41–52
  2. Jump up^ Goldman JM, Melo JV (October 2003). “Chronic myeloid leukemia–advances in biology and new approaches to treatment”. N. Engl. J. Med. 349 (15): 1451–64.doi:10.1056/NEJMra020777. PMID 14534339.
  3. Jump up^ Fausel, C. Targeted chronic myeloid leukemia therapy: Seeking a cure. Am J Health Syst Pharm 64, S9-15 (2007)
  4. Jump up^ Stegmeier F, Warmuth M, Sellers WR, Dorsch M (May 2010). “Targeted cancer therapies in the twenty-first century: lessons from imatinib”. Clin. Pharmacol. Ther. 87(5): 543–52. doi:10.1038/clpt.2009.297. PMID 20237469.
  5. Jump up^ “Novartis fails to patent Glivec (Gleevec) in India”.
  6. Jump up^ Rowley to receive Japan Prize for her role in the development of targeted cancer therapy Eurekalert, Press release, 24 January 2012
  7. Jump up^ Leukemia Drug and Magnet Material Net Japan Prizes by Dennis Normile, Science Insider, 25 January 2012
  8. ^ Jump up to:a b c “FDA Highlights and Prescribing Information for Gleevec(imatinib mesylate)”.
  9. Jump up^ “Prolonged Use of Imatinib in GIST Patients Leads to New FDA Approval”.
  10. Jump up^ “FDA approves Gleevec for children with acute lymphoblastic leukemia”. FDA News Release. US Food and Drug Administration. 25 January 2013. Retrieved 3 April 2013.
  11. Jump up^ Yang FC, Ingram DA, Chen S, Zhu Y, Yuan J, Li X, Yang X, Knowles S, Horn W, Li Y, Zhang S, Yang Y, Vakili ST, Yu M, Burns D, Robertson K, Hutchins G, Parada LF, Clapp DW (October 2008). “Nf1-dependent tumors require a microenvironment containing Nf1+/–and c-kit-dependent bone marrow”. Cell 135 (3): 437–48.doi:10.1016/j.cell.2008.08.041. PMC 2788814. PMID 18984156. Lay summary –Science Daily.
  12. Jump up^ “Gleevec NF1 Trial”. Nfcure.org. Retrieved 2013-04-03.
  13. Jump up^ “GIST in Neurofibromatosis 1”. Gistsupport.org. 2010-05-14. Retrieved 2013-04-03.
  14. Jump up^ “”Pilot Study of Gleevec/Imatinib Mesylate (STI-571, NSC 716051) in Neurofibromatosis (NF1) Patient With Plexiform Neurofibromas (0908-09)” (Suspended)”. Clinicaltrials.gov. Retrieved 2013-04-03.
  15. Jump up^ Droogendijk HJ, Kluin-Nelemans HJ, van Doormaal JJ, Oranje AP, van de Loosdrecht AA, van Daele PL (July 2006). “Imatinib mesylate in the treatment of systemic mastocytosis: a phase II trial”. Cancer 107 (2): 345–51. doi:10.1002/cncr.21996.PMID 16779792.
  16. Jump up^ Tapper EB, Knowles D, Heffron T, Lawrence EC, Csete M (June 2009). “Portopulmonary hypertension: imatinib as a novel treatment and the Emory experience with this condition”. Transplant. Proc. 41 (5): 1969–71.doi:10.1016/j.transproceed.2009.02.100. PMID 19545770.
  17. Jump up^ Boucher P, Gotthardt M, Li WP, Anderson RG, Herz J (April 2003). “LRP: role in vascular wall integrity and protection from atherosclerosis”. Science 300 (5617): 329–32.doi:10.1126/science.1082095. PMID 12690199.
  18. Jump up^ Lassila M, Allen TJ, Cao Z, Thallas V, Jandeleit-Dahm KA, Candido R, Cooper ME (May 2004). “Imatinib attenuates diabetes-associated atherosclerosis”. Arterioscler. Thromb. Vasc. Biol. 24 (5): 935–42. doi:10.1161/01.ATV.0000124105.39900.db.PMID 14988091.
  19. Jump up^ Reeves PM, Bommarius B, Lebeis S, McNulty S, Christensen J, Swimm A, Chahroudi A, Chavan R, Feinberg MB, Veach D, Bornmann W, Sherman M, Kalman D (July 2005). “Disabling poxvirus pathogenesis by inhibition of Abl-family tyrosine kinases”. Nat. Med.11 (7): 731–9. doi:10.1038/nm1265. PMID 15980865.
  20. Jump up^ He G, Luo W, Li P, Remmers C, Netzer WJ, Hendrick J, Bettayeb K, Flajolet M, Gorelick F, Wennogle LP, Greengard P (September 2010). “Gamma-secretase activating protein is a therapeutic target for Alzheimer’s disease”. Nature 467 (7311): 95–8.doi:10.1038/nature09325. PMC 2936959. PMID 20811458.
  21. Jump up^ “Alzheimer’s may start in liver – Health – Alzheimer’s Disease | NBC News”. MSNBC. Retrieved 2013-01-06.
  22. Jump up^ Holmes C, Boche D, Wilkinson D, Yadegarfar G, Hopkins V, Bayer A, Jones RW, Bullock R, Love S, Neal JW, Zotova E, Nicoll JA (July 2008). “Long-term effects of Abeta42 immunisation in Alzheimer’s disease: follow-up of a randomised, placebo-controlled phase I trial”. Lancet 372 (9634): 216–23. doi:10.1016/S0140-6736(08)61075-2. PMID 18640458.
  23. Jump up^ Eliminating Morphine Tolerance – Reformulated Imatinib 23 Feb 2012, 5:00 PST
  24. Jump up^ “GLIVEC Tablets – Summary of Product Characteristics (SPC)”. electronic Medicines Compendium. Novartis Pharmaceuticals UK Ltd.
  25. ^ Jump up to:a b c “Gleevec (imatinib) dosing, indications, interactions, adverse effects, and more”.Medscape Reference. WebMD. Retrieved 24 January 2014.
  26. Jump up^ “Imatinib”. Macmillan Cancer Support. Retrieved 26 December 2012.
  27. ^ Jump up to:a b Haberfeld, H, ed. (2009). Austria-Codex (in German) (2009/2010 ed.). Vienna: Österreichischer Apothekerverlag. ISBN 3-85200-196-X.
  28. Jump up^ Kerkelä R, Grazette L, Yacobi R, Iliescu C, Patten R, Beahm C, Walters B, Shevtsov S, Pesant S, Clubb FJ, Rosenzweig A, Salomon RN, Van Etten RA, Alroy J, Durand JB, Force T (August 2006). “Cardiotoxicity of the cancer therapeutic agent imatinib mesylate”. Nat. Med. 12 (8): 908–16. doi:10.1038/nm1446. PMID 16862153.
  29. Jump up^ Shima H, Tokuyama M, Tanizawa A, Tono C, Hamamoto K, Muramatsu H, Watanabe A, Hotta N, Ito M, Kurosawa H, Kato K, Tsurusawa M, Horibe K, Shimada H (October 2011). “Distinct impact of imatinib on growth at prepubertal and pubertal ages of children with chronic myeloid leukemia”. J. Pediatr. 159 (4): 676–81.doi:10.1016/j.jpeds.2011.03.046. PMID 21592517.
  30. ^ Jump up to:a b c d “GLIVEC (imatinib)” (PDF). TGA eBusiness Services. Novartis Pharmaceuticals Australia Pty Ltd. 21 August 2013. Retrieved 24 January 2014.
  31. Jump up^ PDB 1IEP; Nagar B, Bornmann WG, Pellicena P, Schindler T, Veach DR, Miller WT, Clarkson B, Kuriyan J (August 2002). “Crystal structures of the kinase domain of c-Abl in complex with the small molecule inhibitors PD173955 and imatinib (STI-571)”. Cancer Res. 62 (15): 4236–43. PMID 12154025.
  32. Jump up^ Takimoto CH, Calvo E. “Principles of Oncologic Pharmacotherapy” in Pazdur R, Wagman LD, Camphausen KA, Hoskins WJ (Eds)Cancer Management: A Multidisciplinary Approach. 11 ed. 2008.
  33. Jump up^ Gambacorti-Passerini CB, Gunby RH, Piazza R, Galietta A, Rostagno R, Scapozza L (February 2003). “Molecular mechanisms of resistance to imatinib in Philadelphia-chromosome-positive leukaemias”. Lancet Oncol. 4 (2): 75–85. doi:10.1016/S1470-2045(03)00979-3. PMID 12573349.
  34. Jump up^ Deininger MW, Druker BJ (September 2003). “Specific targeted therapy of chronic myelogenous leukemia with imatinib”. Pharmacol. Rev. 55 (3): 401–23.doi:10.1124/pr.55.3.4. PMID 12869662.
  35. Jump up^ Vigneri P, Wang JY (February 2001). “Induction of apoptosis in chronic myelogenous leukemia cells through nuclear entrapment of BCR-ABL tyrosine kinase”. Nat. Med. 7 (2): 228–34. doi:10.1038/84683. PMID 11175855.
  36. Jump up^ Weisberg E, Manley PW, Cowan-Jacob SW, Hochhaus A, Griffin JD (May 2007). “Second generation inhibitors of BCR-ABL for the treatment of imatinib-resistant chronic myeloid leukaemia”. Nature Reviews Cancer 7 (5): 345–56. doi:10.1038/nrc2126.PMID 17457302.
  37. Jump up^ Scheinfeld N, Schienfeld N (February 2006). “A comprehensive review of imatinib mesylate (Gleevec) for dermatological diseases”. J Drugs Dermatol 5 (2): 117–22.PMID 16485879.
  38. Jump up^ Klopp, T, ed. (2010). Arzneimittel-Interaktionen (in German) (2010/2011 ed.). Arbeitsgemeinschaft für Pharmazeutische Information. ISBN 978-3-85200-207-1.
  39. ^ Jump up to:a b Staff, Innovation.org (a project of the Pharmaceutical Research and Manufacturers of America)The Story of Gleevec
  40. Jump up^ Claudia Dreifus for the New York Times. November 2, 2009 Researcher Behind the Drug Gleevec
  41. ^ Jump up to:a b A Conversation With Brian J. Druker, M.D., Researcher Behind the Drug Gleevecby Claudia Dreifus, The New York Times, 2 November 2009
  42. Jump up^ Gambacorti-Passerini C (2008). “Part I: Milestones in personalised medicine—imatinib”. Lancet Oncology 9 (600): 600. doi:10.1016/S1470-2045(08)70152-9.PMID 18510992.
  43. Jump up^ Druker BJ, Lydon NB (January 2000). “Lessons learned from the development of an abl tyrosine kinase inhibitor for chronic myelogenous leukemia”. J. Clin. Invest. 105 (1): 3–7. doi:10.1172/JCI9083. PMC 382593. PMID 10619854.
  44. ^ Jump up to:a b c U.S. Patent 5,521,184
  45. Jump up^ “Imatinib Patent Family”. Espacenet. 1996. Retrieved 2014-07-23.
  46. ^ Jump up to:a b EP 0564409
  47. Jump up^ Staff, European Medicines Agency, 2004.EMEA Scientific Discussion of Glivec
  48. Jump up^ Note: The Indian patent application, which became the subject of litigation in India that gathered a lot of press, does not appear to be publicly available. However according todocuments produced in the course of that litigation (page 27), “The Appellant’s application under the PCT was substantially on the same invention as had been made in India.”
  49. ^ Jump up to:a b WO 9903854
  50. Jump up^ U.S. Patent 6,894,051
  51. Jump up^ FDA Orange Book; Patent and Exclusivity Search Results from query on Appl No 021588 Product 001 in the OB_Rx list.
  52. Jump up^ Novartis press release, May 10, 2001. [http://www.evaluategroup.com/Universal/View.aspx?type=Story&id=5838 FDA approves Novartis’ unique cancer medication Glivec®
  53. Jump up^ Cohen MH et al. Approval Summary for Imatinib Mesylate Capsules in the Treatment of Chronic Myelogenous Leukemia Clin Cancer Res May 2002 8; 935
  54. Jump up^ Margot J. Fromer for Oncology Times. December 2002. What’s in a Name? Quite a Lot When It Comes to Marketing & Selling New Cancer Drugs
  55. Jump up^ Novartis Press Release. April 30 2001Novartis Oncology Changes Trade Name of Investigational Agent Glivec(TM) to Gleevec(TM) in the United States
  56. Jump up^ Experts in Chronic Myeloid Leukemia. The price of drugs for chronic myeloid leukemia (CML) is a reflection of the unsustainable prices of cancer drugs: from the perspective of a large group of CML experts Blood. 2013 May 30;121(22):4439-42. PMID 23620577
  57. Jump up^ Andrew Pollack for the New York Times, April 25, 2013 Doctors Denounce Cancer Drug Prices of $100,000 a Year
  58. Jump up^ Schiffer CA (July 2007). “BCR-ABL tyrosine kinase inhibitors for chronic myelogenous leukemia”. N. Engl. J. Med. 357 (3): 258–65. doi:10.1056/NEJMct071828.PMID 17634461.
  59. Jump up^ As Pills Treat Cancer, Insurance Lags Behind, By ANDREW POLLACK, New York Times, 14 April 2009
  60. Jump up^ Living With a Formerly fatal Blood Cancer, By JANE E. BRODY, New York Times, 18 January 2010
  61. Jump up^ Patented Medicine Review Board (Canada). Report on New Patented Drugs – Gleevec.
  62. Jump up^ “pharmacychecker.com”. pharmacychecker.com. Retrieved 2013-04-03.
  63. Jump up^ Gardiner Harris and Katie Thomas for the New York Times. April 1 2013 Top court in India rejects Novartis drug patent
  64. Jump up^ Note: The Indian patent application No.1602/MAS/1998 does not appear to be publicly available. However according to the decision of the IPAB on 26 June 2009 (page 27) discussed below, “The Appellant’s application under the PCT was substantially on the same invention as had been made in India.”
  65. Jump up^ Staff, European Medicines Agency, 2004. EMEA Scientific Discussion of Glivec
  66. Jump up^ Indian Supreme Court Decision paragraphs 5-6
  67. Jump up^ Novartis v UoI, para 8-9
  68. ^ Jump up to:a b Shamnad Basheer for Spicy IP March 11, 2006First Mailbox Opposition (Gleevec) Decided in India
  69. Jump up^ Staff, LawyersCollective. September 6, 2011[http://www.lawyerscollective.org/news/archived-news-a-articles/126-novartis-case-background-and-update-supreme-court-of-india-to-recommence-hearing.html Novartis case: background and update – Supreme Court of India to recommence hearing
  70. Jump up^ R. Jai Krishna and Jeanne Whalen for the Wall Street Journal. April 1, 2013Novartis Loses Glivec Patent Battle in India
  71. Jump up^ Intellectual Property Appellate Board decision dated 26 June 2009, p 149
  72. Jump up^ W.P. No.24759 of 2006
  73. Jump up^ “Supreme Court rejects bid by Novartis to patent Glivec”.
  74. Jump up^ Novartis v UoI, Para 191
  75. Jump up^ Novartis v UoI, Para 24-25
  76. Jump up^ “How the Indian judgment will reverberate across the world”.
  77. Jump up^ “Patented drugs must be priced smartly”.
  78. Jump up^ Patent with a purpose, Prof. Shamnad Basheer, Indian Express, 3 April 2013
  79.  Kevin Grogan for PharmaTimes. February 27, 2012 Novartis explains stance over India patent law challenge
  80.  Berne Declaration. May 8, 2007 Short questions and answers about the court case initiated by Novartis in India

External links

Imatinib

Title: Imatinib
CAS Registry Number: 152459-95-5
CAS Name: 4-[(4-Methyl-1-piperazinyl)methyl]-N-[4-methyl-3-[[4-(3-pyridinyl)-2-pyrimidinyl]amino]phenyl]benzamide
Additional Names: N-[5-[4-(4-methylpiperazinomethyl)benzoylamido]-2-methylphenyl]-4-(3-pyridyl)-2-pyrimidineamine
Molecular Formula: C29H31N7O
Molecular Weight: 493.60
Percent Composition: C 70.57%, H 6.33%, N 19.86%, O 3.24%
Literature References: Tyrosine kinase inhibitor; highly specific for BCR-ABL, the enzyme associated with chronic myelogenous leukemia (CML) and certain forms of acute lymphoblastic leukemia (ALL). Also shown to inhibit the transmembrane receptor KIT and platelet-derived growth factor (PDGF) receptors. Prepn: J. Zimmermann, EP 564409; idem, US 5521184 (1993, 1996 both to Ciba-Geigy); idem et al., Bioorg. Med. Chem. Lett. 7, 187 (1997). Structural mechanism of ABL specificity: T. Schindler et al., Science 289, 1938 (2000). Activity vs KIT and PDGF receptor kinases: E. Buchdunger et al., J. Pharmacol. Exp. Ther. 295, 139 (2000). Clinical trial in CML: H. Kantarjian et al., N. Engl. J. Med. 346, 645 (2002); in gastrointestinal stromal tumors related to KIT: G. D. Demetri et al., ibid. 347, 472 (2002). Review of clinical experience: D. G. Savage, K. H. Antman, ibid. 346, 683-693 (2002); and pharmacology: V. K. Pindolia et al., Pharmacotherapy 22, 1249-1265 (2002); and development of therapeutic target: B. J. Druker, Adv. Cancer Res. 91, 1-30 (2004).
Properties: mp 211-213°. pKa1 8.07; pKa2 3.73; pKa3 2.56; pKa4 1.52.
Melting point: mp 211-213°
pKa: pKa1 8.07; pKa2 3.73; pKa3 2.56; pKa4 1.52
 
Derivative Type: Methanesulfonate
CAS Registry Number: 220127-57-1
Manufacturers’ Codes: STI-571; CGP-57148B
Trademarks: Gleevec (Novartis); Glivec (Novartis)
Molecular Formula: C29H31N7O.CH3SO3H
Molecular Weight: 589.71
Percent Composition: C 61.10%, H 5.98%, N 16.63%, O 10.85%, S 5.44%
Literature References: Prepn of crystalline form: J. Zimmermann et al., WO 9903854 (1999 to Novartis).
Properties: Occurs in 2 crystalline modifications. a-form, begins to melt at 226°; b-form, mp 217°. Lipophilic at pH 7.4. Soly in water: >100 g/l (pH 4.2); 49 mg/l (pH 7.4).
Melting point: mp 217°
Therap-Cat: Antineoplastic.
Keywords: Antineoplastic, Tyrosine Kinase Inhibitors,  imatinib mesylate, GGP-57148B, STI-571, CGP-57148 (free base), Gleevec, Glivec, imatinib
IMATINIB BASE
Mp 206 – 207 °C (lit.:1   207 – 210 °C);
1=  1 Y.‐F. Liu, C.‐L. Wang, Y.‐J. Bai, N. Han, J.‐P. Jiao and X.‐L. Qi, Org. Process Res. Dev., 2008, 12, 490.
IR νmax/cm-1 3275.0(w), 2928.5(w),
2796.5(w), 1645.9(m), 1586.0(m), 1575.1(s), 1554.0(m), 1531.5(s), 1510.3(m), 1478.1(m),
1448.9(s), 1416.7(m), 1377.7(m), 1352.2(m), 1334.8(m), 1325.6(m), 1308.8(m), 1290.3(s),
1261.1(m), 1204.3(m), 1164.1(m), 1141.7(m), 1124.6(w), 1102.6(m), 1089.2(w), 1052.0(w),
1024.4(w), 1010.0(m), 992.5(w), 968.3(w), 924.5(w), 886.2(w), 857.9(w), 850.3(w),
807.8(m), 795.7(s), 748.1(m), 703.2(m), 690.1(m), 670.7(m);
δH (d6-DMSO, 600 MHz) =
10.14 (1 H, s, NH), 9.26 (1 H, d, J = 1.5 Hz, 2H-pyridin-3-yl), 8.95 (1 H, s, NH), 8.66 (1 H, dd,
J = 4.8 and 1.2 Hz, 6H-pyridin-3-yl), 8.49 (1 H, d, J = 5.1 Hz, 6H-pyridin-2-amine), 8.46 (1 H,
ddd, J = 7.9, 1.5 and 1.2 Hz, 4H-pyridin-3-yl), 8.06 (1 H, d, J = 1.5 Hz, 3H-2-aminotoluene),
7.89 (2 H, d, J = 8.1 Hz, 2H-benzamide), 7.50 (1 H, dd, J = 7.9 and 4.8 Hz, 5H-pyridin-3-yl),
7.46 (1 H, dd, J = 8.3 and 1.5 Hz, 5H-2-aminotoluene), 7.42 – 7.40 (3 H, m, 3H-benzamide
and 5H-pyridin-2-amine), 7.18 (1 H, d, J = 8.3 Hz, 6H-2-aminotoluene), 3.51 (2 H, s, CH2),
2.50 – 2.20 (8 H, m, piperazine CH2), 2.20 (3 H, s, CCH3), 2.13 (3 H, s, NCH3);
δC (CDCl3,
150 MHz) = 165.42(C), 162.72(C), 160.57(C), 158.99(CH), 151.44(CH), 148.48(CH),
142.52(C), 137.77(C), 136.60(C), 134.92(CH), 133.88(C), 132.66(C), 130.75(CH),
129.28(CH), 127.00(CH), 124.23(C), 123.71(CH), 115.35(CH), 113.19(CH), 108.32(CH),
62.49(CH2), 55.07(CH2), 53.10(CH2), 45.98(CH3), 17.65(CH3);
Rf (MeOH) = 0.09; Rt 3.48,
M+H m/z = 494.2; HRMS calculated for C29H31N7ONa [M + Na]+, 516.2488; found 516.2491.
Inline image 1
1H NMR
Inline image 3
13 C NMR
Inline image 4
WO2006024863A1 * 2 Sep 2005 9 Mar 2006 Cipla Ltd Stable crystal form of imatinib mesylate and process for the preparation thereof
WO2006048890A1 * 20 Oct 2005 11 May 2006 Raja Jyotir Jani Imatinib mesylate crystal form and process for preparation thereof
WO2006054314A1 * 11 Aug 2005 26 May 2006 Natco Pharma Ltd Polymorphic forms of imatinib mesylate
WO2007023182A1 * 24 Aug 2006 1 Mar 2007 Novartis Ag Delta and epsilon crystal forms of imatinib mesylate
WO2007059963A1 * 23 Nov 2006 31 May 2007 Novartis Ag F,g,h,i and k crystal forms of imatinib mesylate
WO2007136510A2 * 27 Apr 2007 29 Nov 2007 Ivax Pharmaceuticals Sro Polymorphic forms of imatinib mesylate and processes for their preparation as well as of amorphous imatinib mesylate and form alpha
WO2008150481A2 * 29 May 2008 11 Dec 2008 Sicor Inc Processes for the preparation of crystalline form beta of imatinib mesylate
WO2010133976A2 24 May 2010 25 Nov 2010 Actavis Group Ptc Ehf Substantially pure imatinib or a pharmaceutically acceptable salt thereof
WO2011023146A1 19 Aug 2010 3 Mar 2011 Zentiva, K.S. Imatinib mesylate polymorphs generated by crystallization in aqueous inorganic salt solutions
WO2011049474A1 21 Oct 2010 28 Apr 2011 Tomasz Kozluk Salts of imatinib with tartaric acids
WO2011095835A1 22 Dec 2010 11 Aug 2011 Actavis Group Ptc Ehf Highly pure imatinib or a pharmaceutically acceptable salt thereof
WO2011099039A1 15 Feb 2011 18 Aug 2011 Reliance Life Sciences Pvt. Ltd. Process for the preparation of alpha form of imatinib mesylate
WO2011108953A1 4 Mar 2011 9 Sep 2011 Tomasz Kozluk PROCESS FOR PREPARATION OF POLYMORPHIC FORM α AND NEW POLYMORPHIC FORM OF IMATINIB MESYLATE ISOLATED IN THAT PROCESS
WO2011114337A1 15 Mar 2010 22 Sep 2011 Natco Pharma Limited Process for the preparation of highly pure crystalline imatinib base
WO2011157450A1 17 Jun 2011 22 Dec 2011 Krka, D. D., Novo Mesto New polymorphic form of imatinib base and preparation of salts thereof
WO2011158255A1 * 14 Jun 2011 22 Dec 2011 Aptuit Laurus Private Limited Process for preparation of stable imatintb mesylate alpha form
WO2012014000A1 * 30 Jul 2010 2 Feb 2012 Narain Singh Awadesh STABLE α-CRYSTAL FORM OF IMATINIB MESYLATE AND PREPARING PROCESS THEREOF
WO2013136141A1 * 22 Aug 2012 19 Sep 2013 Fresenius Kabi Oncology Ltd. An improved process for the preparation of alpha form of imatinib mesylate
CN102321070A * 27 Jul 2011 18 Jan 2012 江苏先声药物研究有限公司 Method for preparing imatinib methylolsulfonate alpha crystal through inverse solvent recrystallization method
CN102321070B 27 Jul 2011 22 May 2013 江苏先声药物研究有限公司 Method for preparing imatinib methylolsulfonate alpha crystal through inverse solvent recrystallization method
CN102633775B 6 Apr 2012 17 Jul 2013 江南大学 Method for preparing alpha-crystal-form imatinib mesylate
DE102007021043B4 * 4 May 2007 8 Apr 2010 Chemagis Ltd. Alpha-Form von Imatinib Mesylat und Verfahren zu seiner Herstellung
EP1988089A1 26 Oct 2007 5 Nov 2008 Sicor, Inc. Imatinib base, and imatinib mesylate and processes for preparation thereof
EP2009008A1 26 Oct 2007 31 Dec 2008 Sicor, Inc. Imatinib base, and imatinib mesylate and processes for preparation thereof
EP2311821A1 27 Apr 2007 20 Apr 2011 Sicor, Inc. Polymorphic form of Imatinib mesylate and processes for its preparation
EP2497464A2 20 Feb 2012 12 Sep 2012 Adamed SP. Z O.O. Pharmaceutical composition of imatinibe methanesulphonate and a process for its manufacture
EP2546248A1 * 23 Nov 2006 16 Jan 2013 Novartis AG Crystal form H of imatinib mesylate
EP2578580A1 * 23 Nov 2006 10 Apr 2013 Novartis AG G, I and K crystal forms of imatinib mesylate
EP2749557A1 * 31 Dec 2012 2 Jul 2014 Deva Holding Anonim Sirketi Process for preparation of alpha polymorph of imatinib mesylate from IPA and THF solvate forms of imatinib mesylate
EP2829538A1 * 27 Apr 2007 28 Jan 2015 Sicor, Inc. Polymorphic form of imatinib mesylate and process for its preparation
US7550591 2 May 2007 23 Jun 2009 Chemagis Ltd. 4-(4-methyl-piperazin-1-ylmethyl-benzoic acid with N-(5- amino-2-methylphenyl)-4-(3pyridyl)-2-pyrimidine-amine in the presence of a coupling reagent such as 2-(5-norborene-2,3- dicarboximido)- 1,1,3,3-tetramethyluronium tetrafluoroborate to produce imatinib, salt formation wtih methanesululfonic acid
US7879860 24 Aug 2006 1 Feb 2011 Novartis Ag Variations in melting points, hygroscopicities, solubilities, flow properties and/or thermodynamic stabilities
US7893076 23 Nov 2006 22 Feb 2011 Novartis Ag Crystalline form F of the methanesulfonic acid addition salt of Imatinib, 4-(4-methylpiperazin-1-ylmethyl)-N-[4-methyl-3-(4-(pyridin-3-yl)pyrimidin-2-ylamino)phenyl]-benzamide; suitable for topical, enteral, for example oral or rectal, or parenteral administration
US7947699 9 Jan 2009 24 May 2011 Actavis Group Ptc Ehf Anhydrous amorphous imatinib mesylate
US7977348 17 Apr 2008 12 Jul 2011 Sicor Inc. Polymorphic forms of imatinib mesylate and processes for preparation of novel crystalline forms as well as amorphous and form α
US8067421 27 Apr 2007 29 Nov 2011 Sicor Inc. Imatinib mesylate solvates with solvents selected from aliphatic alcohols, ethers, dioxolane, nitromethane, and acetic acid, and crystalline imatinib mesylate characterized by data selected from powder XRD patterns and solid state 13C NMR spectra; solubility in gastric juices
US8198289 14 Jan 2011 12 Jun 2012 Novartis Ag Crystal form H imatinib mesylate for pharmaceutical use
US8269003 2 Sep 2005 18 Sep 2012 Cipla Limited Stable crystal form of imatinib mesylate and process for the preparation thereof
US8414918 25 Sep 2008 9 Apr 2013 Teva Pharmaceutical Industries Ltd. Stable imatinib compositions
US8507515 15 Jul 2011 13 Aug 2013 Novartis Ag Crystalline form G of imatinib mesylate
US8592440 15 Jul 2011 26 Nov 2013 Novartis Ag Crystalline form I of imatinib mesylate
US8633213 15 May 2012 21 Jan 2014 Novartis Ag Crystalline form F of imatinib mesylate
US8846706 15 Jul 2011 30 Sep 2014 Novartis Ag Crystalline form K of imatinib mesylate
USRE43932 21 Sep 2011 15 Jan 2013 Novartis Ag Crystal modification of a N-phenyl-2-pyrimidineamine derivative, processes for its manufacture and its use

—

ANTHONY MELVIN CRASTO

THANKS AND REGARD’S
DR ANTHONY MELVIN CRASTO Ph.D


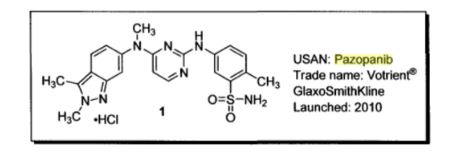
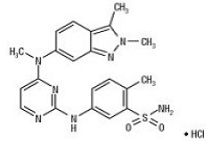
Pazopanib, パゾパニブ塩酸塩 , Пазопаниба Гидрохлорид


Pazopanib3Dan.gif

STR1

Pazopanib

パゾパニブ塩酸塩

Пазопаниба Гидрохлорид

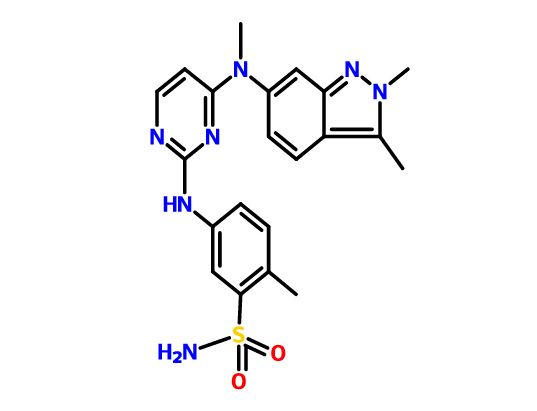
5-[[4-[(2,3-Dimethyl-2H-indazol-6-yl)methylamino]-2-pyrimidinyl]amino]-2-methylbenzolsulfonamide

Pazopanib is a small molecule inhibitor of multiple protein tyrosine kinases with potential antineoplastic activity. It is developed by GlaxoSmithKline and was FDA approved on October 19, 2009.

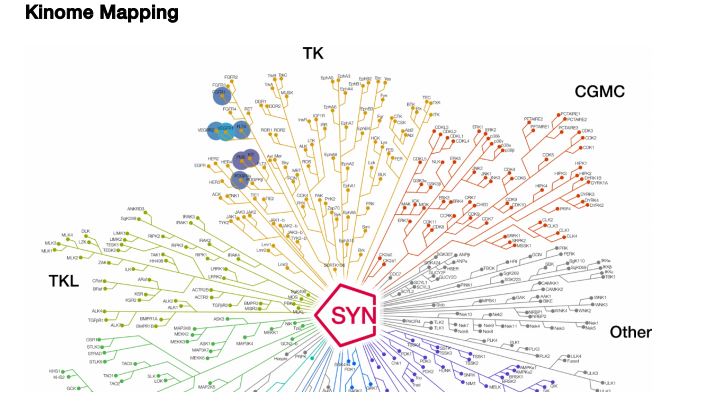
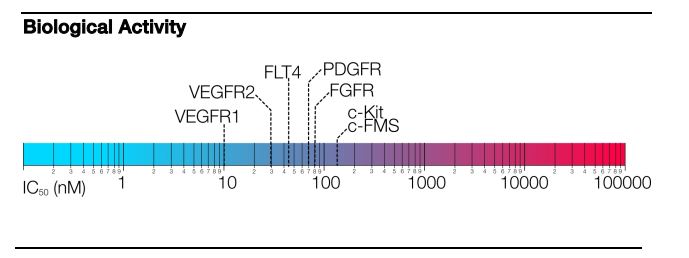
Pazopanib is a potent and selective multi-targeted receptor tyrosine kinase inhibitor of VEGFR-1, VEGFR-2, VEGFR-3,
PDGFR-a/b, and c-kit that blocks tumor growth and inhibits angiogenesis. It was approved for renal cell carcinoma by the U.S. Food  and Drug Administration in 2009 and is marketed under the trade name Votrient by the drug’s manufacturer, GlaxoSmithKline.

GW 786034

M.Wt: 437.53
C21H23N7O2S

Pazopanib CAS No.: 444731-52-6

CAS No.: 635702-64-6 (PAZOPANIB HYDROCHLORIDE)
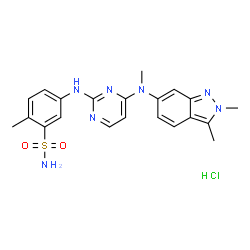
ChemSpider 2D Image | Pazopanib Hydrochloride | C21H24ClN7O2S

Pazopanib Hydrochloride

CAS No.: 635702-64-6 (PAZOPANIB HYDROCHLORIDE)

  • MFC21H24ClN7O2S
  • MW473.979
GW786034;Votrient;Armala;GW 786034;GW-786034
GW786034GW786034, VOTRIENT
5-({4-[(2,3-Dimethyl-2H-indazol-6-yl)(methyl)amino]-2-pyrimidinyl}amino)-2-methylbenzenesulfonamide hydrochloride (1:1)
Antineoplastic; Tyrosine Kinase Inhibitors, Protein Kinase Inhibitors; Renal Cell Carcinoma Therpay; Soft Tissue Sarcoma Therapy
パゾパニブ塩酸塩
Pazopanib Hydrochloride

C21H23N7O2S▪HCl : 473.98
[635702-64-6]

Pazopanib (trade name Votrient) is a potent and selective multi-targeted receptor tyrosine kinase inhibitor that blocks tumour growth and inhibits angiogenesis. It has been approved for renal cell carcinoma and soft tissue sarcoma by numerous regulatory administrations worldwide.[2][3][4][5]

Pazopanib (Votrient®; GlaxoSmithKline, Brentford, U.K.)  is currently approved by the U.S. Food and Drug Administration (FDA) and the European Medicines Agency for the treatment of patients with metastatic renal cell carcinoma (mRCC)

Medical uses

It is approved by numerous regulatory administrations worldwide (including the FDA (19 October 2009), EMA (14 June 2010), MHRA(14 June 2010) and TGA (30 June 2010)) for use as a treatment for advanced/metastatic renal cell carcinoma and advanced soft tissue sarcomas.[1][2][3][4][5] In Australia and New Zealand, it is subsidised under the PBS and by Pharmac respectively, under a number of conditions, including:[6][7]

  • The medication is used to treat clear cell variant renal cell carcinoma.
  • The treatment phase is continuing treatment beyond 3-months.
  • The patient has been issued an authority prescription for pazopanib
  • The patient must have stable or responding disease according to the Response Evaluation Criteria In Solid Tumours (RECIST)
  • This treatment must be the sole tyrosine kinase inhibitor subsidised for this condition.

It has also demonstrated initial therapeutic properties in patients with ovarian and non-small cell lung cancer,[8] though plans to apply to the EMA for a variation to include advanced ovarian cancer have been withdrawn and a license will not be sought in any country.[9][10]

Pazopanib

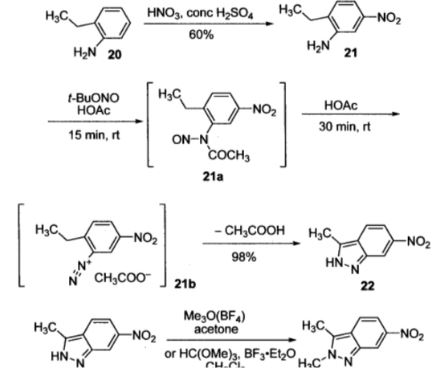
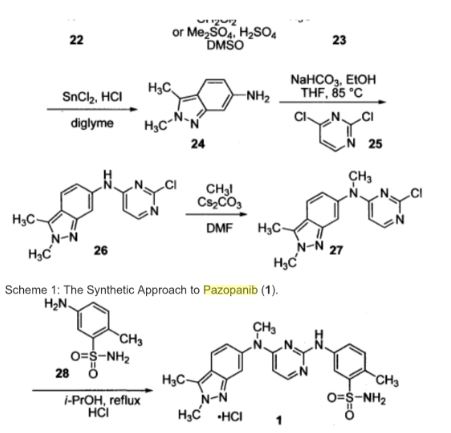
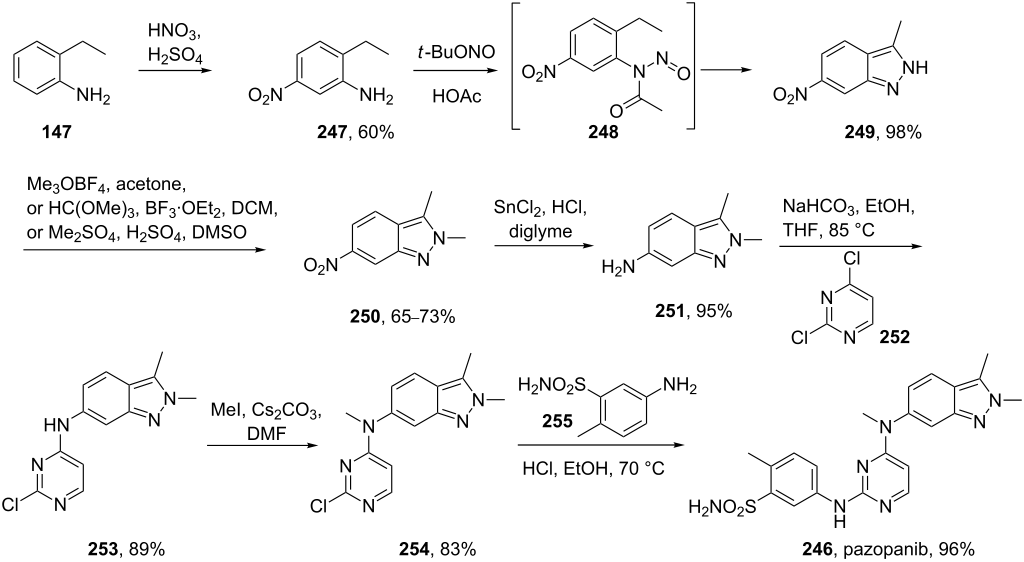
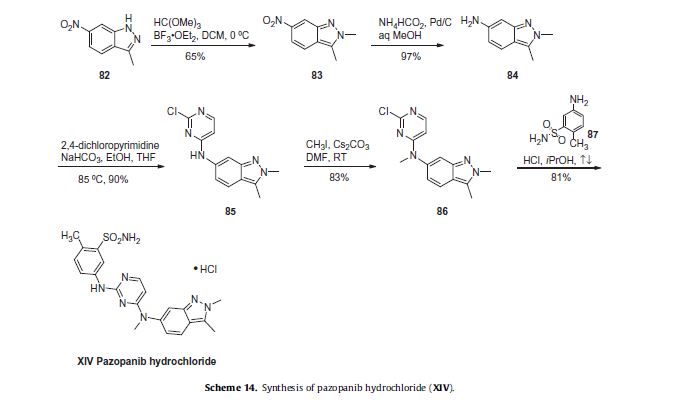
SYNTHESIS

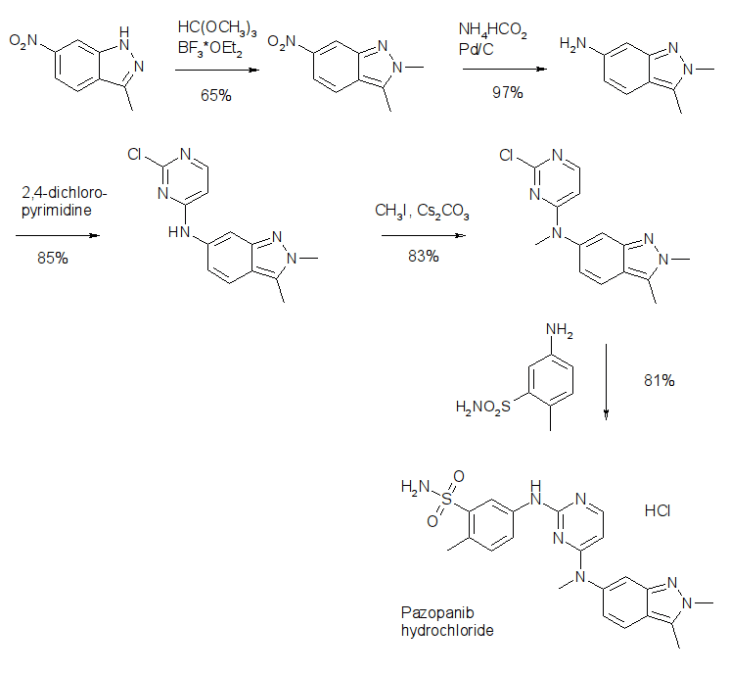
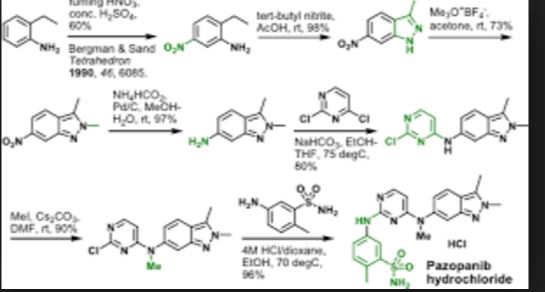
Pazopanib hydrochloride drug substance is manufactured by Glaxo Wellcome Manufacturing Pte. Limited, Jurong, Singapore

NDA 22-465 was submitted by GlaxoSmithKline (GSK) for VOTRIENT™ (pazopanib), an immediate release tablet for oral administration containing either 200 mg or 400 mg of pazopanib free base (GW786034X) as the hydrochloride salt (GW786034B). Pazopanib is a new molecular entity and is submitted for review pursuant to Section 505(b)(1) of the Food, Drug and Cosmetic Act. Reference is made to one active Investigational New Drug application, IND 65,747.

Quality by Design (QbD) approach and risk management to increase their understanding of the process and drug substance properties. A number of Critical Quality Attributes (CQAs) were identified. These are: Identity by IR, Chloride Identity, Crystalline Form, Content by HPLC, Drug-related Impurities (including named impurities and genotoxic (b) (4) (b) (4) (b) (4) (b) (4) Executive Summary Section CMC Review #1 Page 9 of 262 CMC REVIEW OF NDA 22-465 impurities content), Residue on Ignition, Particle Size, Residual Solvents, Water Content by Karl Fischer, Description, Pd Content, and Heavy Metals.

CQA are mainly controlled by controlling starting material attributes, intermediate attributes (e.g. specifications of GW786034 quality process parameters, and by following the manufacturing process. A risk based method (e.g. failure mode and effects analysis (FMEA)) was used to identify the Quality Critical Process Parameters (QCPPs), Quality Process Parameters (QPPs), CQAs and Quality Attributes (QAs) for the pazopanib hydrochloride manufacturing process. The inputs to the FMEA were knowledge gained through the work to develop the impurity fate map, spiking and purging studies, the Design of Experiments (DOE) work to establish potential QCPPs/QPPs, one factor at a time experiments to establish parameter proven acceptable ranges, and 6 years of plant experience preparing over batches of pazopanib hydrochloride in 3 plants throughout the GSK network. It was concluded from this risk assessment that there are no QCPP but a few QPP in the drug substance (DS) manufacturing process. Stages 1 and 2 had no QPP, the few were only present in stages 3 and 4. All the QPP were scale invariant. A combination of multivariate DOE and univariate experimentation was used to determine the Proven acceptable Ranges (PAR) for the variables. The risks for combining univariate and multivariate experimentation were found to be minimal, on the basis of outcome from the robustness study. For this study, all the process parameters were all set at the lower limit of the PARs to create a worst-case scenario for impurity purging. Neither new impurities nor elevated levels of known impurities were detected. This data demonstrated that multivariate interactions will not lead to elevated levels of impurities.

Drug Product Pazopanib Tablets, 200 mg and 400 mg are film-coated IR oral tablets. The two strengths contain 216.7 mg and 433.4 mg pazopanib hydrochloride, respectively, which are equivalent to 200 mg and 400 mg pazopanib (free base), respectively. Excipients in the tablet core are: microcrystalline cellulose, sodium starch glycolate, povidone, and magnesium stearate. Pazopanib Tablets, 200 mg are modified capsule-shaped, gray film-coated tablets, one side plain and the opposite side debossed with an identifying code of ‘GS JT’. Pazopanib Tablets, 400 mg are modified capsule-shaped, yellow film-coated tablets, one side plain and the opposite side debossed with an identifying code of ‘GS UHL’. The tablets are manufactured at Glaxo Operations UK Limited, Priory Street, Ware, Hertfordshire SG12 0DJ, United Kingdom. Primary packaging of tablets will be performed by either Glaxo Operations UK Limited, Priory Street, Ware, Hertfordshire SG12 0DJ, United Kingdom or GlaxoSmithKline Inc, 1011 North Arendell Avenue, Zebulon, North Carolina 27597, USA.

Pazopanib hydrochloride is a new molecular entity of Biopharmaceutics Classification System (BCS) Class 2 (poor solubility, high permeability) and a crystalline solid. Its solubility in pH 1.1 is 0.65 mg/mL. The conjugate acids of the basic nitrogens have the following acidity constants: pKa – pK1 = 2.1 (indazole), pK2 = 6.4 (pyrimidine), pK3 = 10.2 (sulfonamide).

STR1

“Synthetic approaches to the 2009 new drugs”
Kevin K.-C. Liua, Subas M. Sakyab, Christopher J. O’Donnellb, Andrew C. Flickb, Jin Lic,
Bioorganic & Medicinal Chemistry, Volume 19, Issue 3, Pages 1136–1154

“An overview of the key routes to the best selling 5-membered ring heterocyclic pharmaceuticals”., Beilstein J. Org. Chem., 2011, 7, 442–495.

STR1 STR2

PATENT

https://www.google.com/patents/WO2015068175A2?cl=en

Pazopanib is marketed as hydrochloride salt by Glaxoshiithkline under the trade name VOTRIENT® is tyrosine kinase inhibitor and indicated for the treatment of patients with advanced renal cell carcinoma (RCC) and treatment of patients with advanced soft tissue sarcoma (STS) who have received prior chemotherapy.

U.S. Patent No (s). US 7105530 (“the ‘530 patent”), US7262203 (“the ‘203 patent”) and US8114885 (“the ‘885 patent”) discloses a variety of pyrimidineamines and their derivatives such as Pazopanib, processes for their preparation, pharmaceutical compositions comprising the derivatives, and method of use thereof.

The process disclosed in the ‘530 patent is schematically represented as follows:

STR1

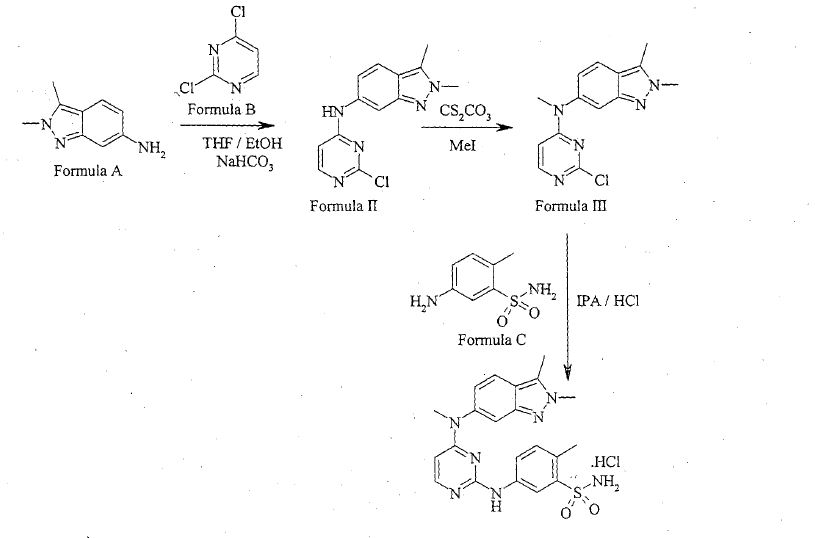
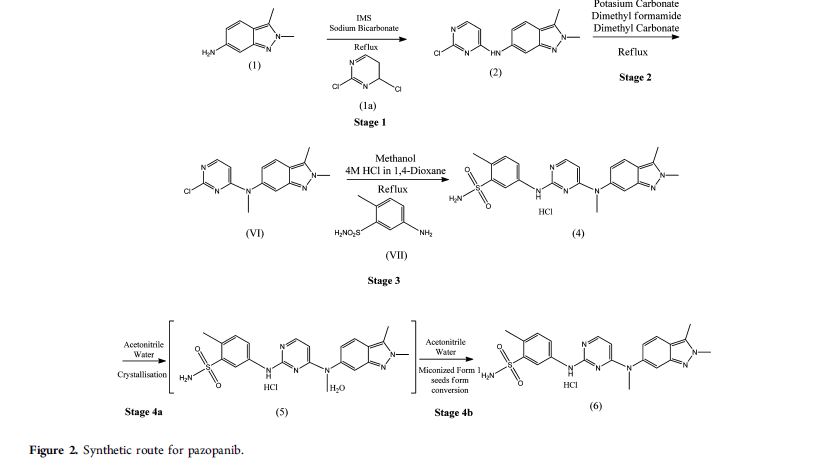
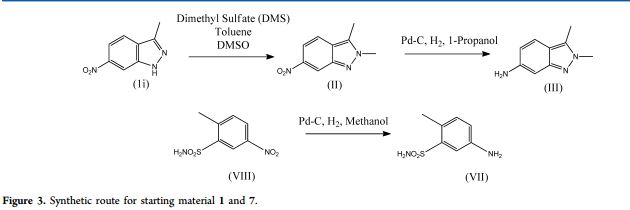
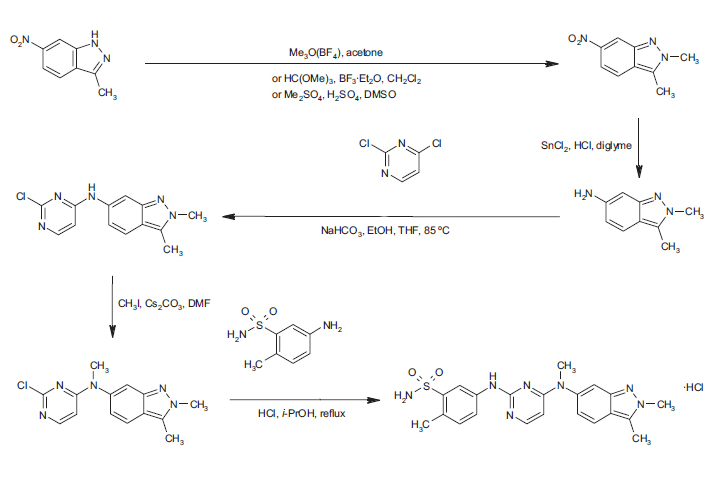
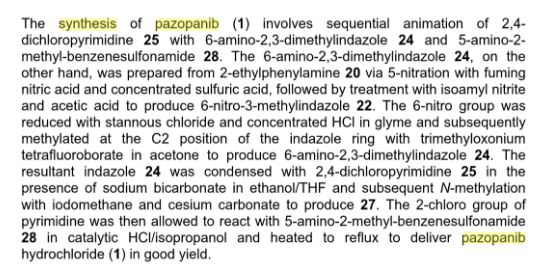
Patent publication No. WO 2011/050159 (“the Ί59 publication”) disclosed process for preparation of Pazopanib hydrochloride, which involves condensation of 2,3-dimethyl-2H-indazol-6-amine of Formula A and 2,4-dicMoropyrimidine of Formula B in a solvent like industrial methylated sprit and specific reaction conditions like, in presence of a base, sodium bicarbonate having a particle size distribution of > 250μηι or 50 to 150μηι selected to ensure that the pH of the reaction mixture is less than 7 for the reaction time period not more than 300 min to obtain N-(2-c oropyrimidin-4-yl)- 2,3-dimethyl-2H-indazol-6-amine of Formula II. The compound of Formula Π was methylated in presence of a methylating agent in an organic solvent like dimethylformamide by using specific reaction conditions like, in presence of a base i.e. Potassium carbonate having a particle size distribution D99 of > 300μηι or D99 of < 200μηι selected to ensure that the reaction time needs to reduce the starting material to less than 2% in less than 8 his to obtain N-(2-cMoropyrimidin-4-yl)-N-2,3-trimemyl-2H-mdazol-6-amine of Formula III. The resultant methylated compound was condensed with 5-amino-2-methylbenzenesulfonamide of Formula C in presence of 4M HC1 and methanol to yield Pazopanib hydrochloride.

WO Ί59 publication disclosed that use of sodium bicarbonate with specific particle size distribution of > 250μπι or 50 to 150μηι is key element in condensation of compound of Formula A and Formula B to niinimize the formation of Impurity of Formula 1 within

WO Ί59 publication also disclosed that use of potassium carbonate with specific particle size distribution D99 of > 300μπι or D99 of < 200μηι is key element in methylation of compound of Formula II to reduce the formation of Impurities of Formula 2, Formula 3 and Formula 4 within the range of about 0.05-3%.

Patent publication No. WO 2012/073254 (“the ‘254 publication”) disclosed a process for preparation of pazopanib hydrochloride, which involves condensation of 2,4-dicMoropyrimidine of Formula B with 5-amino-2-methylbenzenesulfonamide of Formula C in presence of a base like sodium bicarbonate and a solvent like ethanol to yield 5-(4-chloropyrimidm-2-yl-ammo)-2-memylbenzenesulfonamide The resultant compound was condensed with N-2,3-1rimethyl-2H-indazole-6-amine of Formula D in an alcoholic solvent like ethanol. WO ‘254 publication also discloses process for purification of pazopanib hydrochloride from alcoholic solvent and water. The process disclosed in the ‘254 publication is schematically represented as follows:

Patent publication No. IN 2505/CHE/2011 disclosed a process for preparation of pazopanib, which involves condensation of 2,3-dimethyl-2H-indazol-6-amine of Formula A and 2,4-dichloropyrimidine of Formula B in presence of sodium bicarbonate and a phase transfer catalyst like tetrabutyl ammonium bromide in a solvent like methanol to obtain N-(2-chloropyrimidin-4-yl)-2,3 -dimethyl -2H-indazol-6-amine of Formula II. The resultant compound was methylated in presence of methyl iodide, potassium carbonate in a solvent like dimethylformamide to obtain compound of Formula III. The obtained Formula III was condensed with 5-amino-2-methylbenzenesulfonamide of Formula C in presence of dimethylformamide and concentrated HC1 to yield pazopanib hydrochloride.

Patent publication No. CN 103373989 (“the ‘989 publication”) disclosed a process for preparation of Pazopanib intermediate of Formula III by condensation of N-2,3-trimethyl-2H-indazole-6-amine of Formula D with 2,4-dicWoropyrimidine of Formula B in

Patent publication No. WO 2014/97152 (“the Ί52 publication”) disclosed a process for preparation of Pazopanib hydrochloride starting from 2,3-dimethyl-6-nitro-2H-indazole.

The processes for preparation of pazopanib described in the above literature have certain drawbacks as it involves: a) use of specific predefined particles of bases like sodium bicarbonate and potassium carbonate, which involves additional process steps like milling, grinding etc, b) use of expensive phase transfer catalysts and c) multiple steps making the process quite expensive, particularly on large scale.

European Medicines Agency (EMA) public assessment report disclosed that pazopanib hydrochloride is a white to slightly yellow, non-hygroscopic, crystalline substance and the manufacturing process consistently produces pazopanib hydrochloride Form 1. However, the EMEA does not describe any particular characterization data for the disclosed polymorph Form 1.

PCT Publication No. WO 2011/058179 (“the Ί79 publication”) discloses pazopanib base crystalline Forms such as Form-I and Form-II and a process for its preparation; also disclosed characterization data of Form-I and Form-II by XRD, IR and melting point. –

PCT Publication No. WO 2011/069053 (“the ‘053 publication”) discloses crystalline pazopanib base and crystalline pazopanib hydrochloride Forms such as Form-II, Form-Ill, Form-TV, Form-V, Form- VI, Form- VIII, Form-IX, Form-X, Form-XI, Form-XII, Form-XIII, Form-A, Form-G and also discloses crystalline Pazopanib dihydrochloride Forms such as Form-I, Form-XIV, Form-XV. The crystalline Forms reported in the PCT publication characterized by its XRD pattern.

IN Publication No. 3023/CHE/2010 discloses crystalline pazopanib dihydrochloride Form-I and crystalline pazopanib mono hydrochloride, process for it preparation and characterization by XRD of the same.

IN Publication No. 1535/CHE/2012 discloses crystalline pazopanib hydrochloride Form-SP and a process for its preparation; also disclosed characterization data, of Form-SP by XRD, DSC and TR.

PCT Publication No (s): WO 2007/143483, WO 2007/064753, WO 2006/20564 and WO 2005/105094 as well as US Publication No. US 2008/0293691 disclose anhydrous and hydrated Forms of pazopanib hydrochloride and their process for preparation thereof. ‘

IP. Com journal disclosure Number IPCOM000207426D discloses crystalline Form of pazopanib hydrochloride Form-R, which is characterized by XRD pattern.

Further, IP.Com journal disclosure Number IPCOM000193076D discloses crystalline Forms of N-(2-cUoropyrirnidin-4-yl)-N-2,3-trimethyl-2H-indazol-6-amine of

Formula III such as Form I and Form II along with characteristic data of XRD pattern

PATENT

https://www.google.com/patents/WO2012073254A1?cl=en

Examples

Example 1:

Preparation of 5-(4-chloropyrimidin-2ylamino)-2-methyIbenzenesulfonamide

To a mixture of 5-amino-2-methylbenzenesulfonamide (20 gm) in ethanol (208 ml) and tetrahydrofuran (52 ml) was added 2,4-dichloropryrimidine (44 gm) and sodium bicarbonate (36 gm) at room temperature. The contents were heated to 70 to 75°C and maintained for 13 hours. The reaction mass was then cooled to 10°C and maintained for 2 hours. The reaction mass was filtered and the solvent was distilled off under vacuum at below 50 to 55°C to obtain a residual mass. To the residual mass was added ethyl acetate (100 ml) and stirred for 1 hour, filtered. The solid obtained was dried to give 15.5 gm of 5-(4-chloropyrimidin-2ylamino)-2-methylbenzenesulfonamide. Example 2:

Preparation of N,2,3-trimethyI-2H-indazol-6-amine

Sodium methoxide (19 gm) was dissolved in methanol (610 ml) and then added 2,3-dimethyl-2H-indazol-6-amine (13 gm). The reaction mixture was stirred for 15 minutes and then added paraformaldehyde (3.9 gm). The contents were heated to 60°C and stirred for 10 hours. The reaction mass was then cooled to room temperature and maintained for 4 hours 30 minutes. Sodium borohydride (2.8 gm) was added to the reaction mass slowly at room temperature and then heated to reflux. The reaction mass was maintained for 2 hours at reflux and then cooled to room temperature. The reaction mass was stirred for 14 hours at room temperature and then added sodium hydroxide solution (1M, 100 ml). The pH of the reaction mass was adjusted to 8.0 to 8.5 with hydrochloric acid solution (40 ml) and then added ethyl acetate (400 ml). Then the layers were separated and the aqueous layer was extracted with ethyl acetate. The organic layer was dried with sodium sulfate and treated with carbon. The combined organic layers were washed with sodium chloride solution and dried with sodium sulfate. The organic layer was treated with carbon and filtered through hi-flow bed. The solvent was distilled off under vacuum at below 50°C to obtain a residual mass. To the residual mass was added diisopropyl ether (75 ml) and stirred for 1 hour, filtered. The solid obtained was dried to give 10 gm of N,2,3-trimethyl-2H-indazol-6-amine.

Example 3:

Preparation of pazopanib hydrochloride

5-(4-Chloropyrimidin-2ylamino)-2-methylbenzenesulfonamide (17 gm) as obtained in example 1, N,2,3-trimethyl-2H-indazol-6-amine (10 gm) as obtained in example 2 and ethanol (166 ml) were added at room temperature and then heated to reflux. The reaction mass was maintained for 3 hours at reflux and then added concentrated hydrochloric acid (1 ml). The reaction mass was maintained for 10 hours at reflux and then cooled to room temperature. The separated solid was filtered and dried to obtain 17 gm of pazopanib hydrochloride (HPLC Purity: 97.5%). Example 4:

Purification of pazopanib hydrochloride

Pazopanib hydrochloride (5 gm; HPLC Purity: 97.5%) as obtained in example 3 was dissolved in a mixture of methanol (100 ml) and water (10 ml) at room temperature and then heated to reflux. The reaction mass was maintained for 30 minutes at reflux and filtered. The filtrate obtained was cooled to room temperature and maintained for 2 hours at room temperature. The solid obtained was collected by filtration and dried to obtain 3.5 gm of pazopanib hydrochloride (HPLC Purity: 99.9%). Example 5:

Purification of pazopanib hydrochloride

Pazopanib hydrochloride (22 gm; HPLC Purity: 98%), methanol (528 ml), water (55 ml) and concentrated hydrochloric acid (0.2 ml) were added at room temperature. The contents were heated to reflux and maintained for 30 minutes, filtered. Take the filtrate and the solvent was distilled off under vacuum to obtain a residual mass. The residual mass was then cooled to room temperature and stirred for 30 minutes at room temperature. The contents were further cooled to 0 to 5°C, stirred for 1 hour and filtered. The solid obtained was dried to give 19 gm of pazopanib hydrochloride (HPLC Purity: 99.85%).

Example 6:

Purification of pazopanib hydrochloride

Pazopanib hydrochloride (10 gm; HPLC Purity: 96%), methanol (250 ml), water (25 ml) and concentrated hydrochloric acid (0.1 ml) were added at room temperature. The contents were heated to reflux and maintained for 30 minutes, filtered. The filtrate obtained was then cooled to room temperature and stirred for 30 minutes at room temperature. The contents further cooled to 0 to 10°C and stirred for 1 hour. The separated solid was filtered and dried to obtain 6.6 gm of pazopanib hydrochloride (HPLC Purity: 99.8%).

Example 7: Purification of pazopanib hydrochloride

Pazopanib hydrochloride (22 gm; HPLC Purity: 97%) was dissolved in a mixture of isopropanol (132 ml) and water (20 ml) at room temperature and then heated to reflux. The reaction mass was maintained for 1 hour at reflux and then cooled to room temperature. The reaction mass was stirred for 1 hour at room temperature and filtered. The solid obtained was dried to give 18 gm of pazopanib hydrochloride (HPLC Purity: 99.8%).

Paper

http://pubs.acs.org/doi/full/10.1021/op400139z

Assessment of Predictivity of Semiquantitative Risk Assessment Tool: Pazopanib Hydrochloride Genotoxic Impurities

† GlaxoSmithKline, Park Road, Ware, Hertfordshire, United Kingdom SG12 0DP
‡ GlaxoSmithKline, 709 Swedeland Road, King of Prussia, Pennsylvania 19406, United States
Org. Process Res. Dev., 2013, 17 (8), pp 1036–1041
DOI: 10.1021/op400139z
Publication Date (Web): July 02, 2013
Copyright © 2013 American Chemical Society

Abstract

Abstract Image

The recently developed semiquantitative assessment tool for the evaluation of carryover potential of mutagenic impurities (MIs) into the final API was applied to the five identified MIs within pazopanib hydrochloride (dimethyl sulfate (DMS) and compounds II, III, VI, and VIII). The theoretical and predicted purge factors were compared. The tool accurately predicted the purging capacity for the most reactive MI, DMS, giving a theoretical purge factor of 30000 versus an actual value of 29411 (for spiking at stage 1). For the other less reactive MIs, both measured and predicted values agreed reasonably well, and the high values for the purging factors were indicative of an effective process capability that could significantly reduce observed MI levels. The only exception was for compound VI, where although the measured and theoretical purge factors were in agreement, they were significantly lower (<200) than for the other MIs. In this case, a strategy was implemented including a requirement for control of this MI on API specification. The purge-factor assessment tool has the potential to play a key role in GRA (genotoxic risk assessment) processes and subsequent regulatory submissions. This tool could provide regulators with additional confidence to accept these purging arguments without resorting to testing. This could potentially significantly reduce the analytical testing burden for early clinical candidates.

STR1

STR1

PATENT

WO 2011058179

The compound 5-(4-(N-(2,3-dimethyl-2H-indazole-6-yl)-N-methylamino)pyrimidine-2- ylamino)-2-methylbenzenesulfonamide, also known as Pazopanib, is useful in the treatment of disorders associated with inappropriate or pathological angiogenesis, such as cancer, in mammals. Pazopanib has the following formula (I):

H CH,

Figure imgf000002_0001

NH2

In WO 02/059110 the preparation of 5-(4-(N-(2,3-dimethyl-2H-indazole-6-yl)-N- methylamino)pyrimidine-2-ylamino)-2-methylbenzenesulfonamide hydrochloride as well as the uses of this compound have been disclosed. In particular, this compound is an inhibitor of tyrosine kinase enzymes, namely vascular endothelial growth factor receptors, and can be used for the treatment and/or prevention of diseases which are associated with tyrosine kinase enzymes such as vascular endothelial growth factor receptors, such as cancer, particularly breast cancer and colon cancer.

Alternative methods for the preparation of 5-(4-(N-(2,3-dimethyl-2H-indazole-6-yl)-N- methylamino)pyrimidine-2-ylamino)-2-methylbenzenesulfonamide hydrochloride are disclosed in WO 03/106416.

In WO 2007/064752 the use of Pazopanib for the treatment of age related macula degeneration is disclosed. WO 2007/064753 further discloses Pazopanib for the treatment of various types of cancer, e.g. brain cancer, glioblastoma multiforme, neuroendocrine cancer, prostate cancer, myeloma, lung cancer, liver cancer, gallbladder cancer or skin cancer.

Typically Pazopanib is administered orally, as this route provides great comfort and convenience of dosing. Although the hydrochloride form of Pazopanib is known in the art, as described above, this form is not optimal in regard to bioavailability, inter-patient variability, and safety. Further, the known form of Pazopanib hydrochloride is not optimal with regard to mechanical and chemical stability, which is in particular necessary for manufacturing tablets, as well as not optimal in regard to flow properties, compressibility, dissolution rate. Additionally, it is at least to some extent hygroscopic and shows electrostatic charging. These properties constitute disadvantages in the preparation of pharmaceutical compositions

PAPER

Synthesis and biological evaluation of novel pazopanib derivatives as antitumor agents


Abstract

A series of novel pazopanib derivatives, 7a–m, were designed and synthesized by modification of terminal benzene and indazole rings in pazopanib. The structures of all the synthesized compounds were confirmed by 1H NMR and MS. Their inhibitory activity against VEGFR-2, PDGFR-α and c-kit tyrosine kinases were evaluated. All the compounds exhibited definite kinase inhibition, in which compound 7l was most potent with IC50 values of 12 nM against VEGFR-2. Furthermore, compounds 7c, 7d and 7mdemonstrated comparable inhibitory activity against three tyrosine kinases to pazopanib, and compound 7f showed superior inhibitory effects than that of pazopanib.

Chemical structure of pazopanib.

Figure 1.

Chemical structure of pazopanib.

Patent

https://www.google.co.in/patents/WO2002059110A1?cl=en

Example 69

5-({4-[(2,3-dimethyl-2r/-indazol-6-yl)(methyl)amino]pyrimidin-2-yl}amino)-2- methylbenzenesulfonamide

Figure imgf000096_0002

To a solution of Intermediate Example 13 (200 mg, 0.695 mmol) and 5-amino-2- ethylbenzenesulfonamide (129.4 mg, 0.695 mmol) in isopropanol (6 ml) was added 4 drops of cone. HCI. The mixture was heated to reflux overnight. The mixture was cooled to rt and diluted with ether (6 ml). Precipitate was collected via filtration and washed with ether. HCI salt of 5-({4-[(2,3-dimethyl-2H-indazol-6-yl)(methyl)amino]-pyrimidin-2- yl}amino)-2-methylbenzenesulfonamide was isolated as an off-white solid. Y\ NMR (400 MHz, deDMSO+NaHCOa) δ 9.50 (br s, 1 H), 8.55 (br s, 1 H), 7.81 (d, J = 6.2 Hz, 1 H), 7.75 (d, J = 8.7 Hz, 1 H), 7.69 (m, 1 H), 7.43 (s, 1 H), 7.23 (s, 2H), 7.15 (d, J = 8.4 Hz, 1 H), 6.86 (m, 1 H), 5.74 (d, J = 6.1 Hz, 1 H), 4.04 (s, 3H), 3.48 (s, 3H), 2.61 (s, 3H), 2.48 (s, 3H). MS (ES+, m/z) 438 (M+H).

Example 13

Preparation of Λ/-(2-chloropyrimidin-4-yl)-Λ/,2,3-trimethyl-2r/-indazol-6-amine

Figure imgf000061_0001

To a stirred solution of the Intermediate 12 (7.37 g) in DMF (50 ml) was added CS2CO3 (7.44 g, 2 eqv.) and Mel (1.84 ml, 1.1 eqv.) at room temperature. Mixture was stirred at rt for overnight The reaction mixture was poured into ice-water bath, and the precipitate was collected via filtration and washed with water. The precipitate was air- dried to afford Λ/-(2-chloropyrimidin-4-yl)-Λ/,2,3-trimethyl-2r/-indazol-6-amine as an off-white solid (6.43 g, 83%). ‘H NMR (400 MHz, dsDMSO) δ 7.94 (d, J = 6.0 Hz, 1 H), 7.80 (d, J = 7.0 Hz, 1 H), 7.50 (d, J = 1.0 Hz, 1 H), 6.88 (m, 1 H), 6.24 (d, J = 6.2 Hz, 1 H), 4.06 (s, 3H), 3.42 (s, 3H), 2.62 (s, 3H). MS (ES+, m/z) 288 (M+H).

Intermediate Example 12 Preparation of Λ/-(2-chloropyrimidin-4-yl)-2,3-dimethyl-2H-indazol-6-amine

Figure imgf000060_0001

to a stirred solution of Intermediate Example 11 (2.97 g, .015 mol) and NaHCOs (5.05 g, .06 mol) in THF (15 mL) and ethanol (60 mL) was added 2,4-dichloropyrimidine (6.70 g, .045 mol) at room temperature. After the reaction was stirred for four hours at 85 °C, the suspension was cooled to rt, filtered and washed thoroughly with ethyl acetate. The filtrate was concentrated under reduced pressure, and the resulting solid was triturated with ethyl acetate to yield 3.84 g (89 % yield) of Λ/-(2-chloropyrimidin-4-yl)- 2,3-dimethyl-2tf-indazol-6-amine. 1H NMR (400 MHz, deDMSO) δ 7.28 (d, J = 9.0 Hz, 1 H), 6.42 (d, J = 8.8 Hz, 1 H), 6.37 (s, 1 H), 5.18 (br s, 1 H), 3.84 (s, 3H), 2.43 (s, 3H). MS (ES+, m/z) 274 (M+H).

Intermediate Example 11

Preparation of 2,3-dimethyl-2r/-indazol-6-amine

Figure imgf000059_0002

To a stirred solution of 18.5 g (0.11 mol) of 3-methyl-6-nitro- 7W-indazole in 350 ml acetone, at room temperature, was added 20 g (0.14 mol) of trimethyloxonium tetraflouroborate. After the solution was allowed to stir under argon for 3 hours, the solvent was removed under reduced pressure. To the resulting solid was added saturated aqueous NaHC03 (600 ml) and a 4:1 mixture of chloroform-isopropanol (200 ml), and the mixture was agitated and the layers were separated. The aqueous phase was washed with additional chloroform: isopropanol (4 x 200 ml) and the combined organic phase was dried (Na2S04). Filtration and removal of solvent gave a tan solid. The solid was washed with ether (200 ml) to afford 2,3-dimethyl-6-nitro-2r/-indazole as a yellow solid (15.85 g, 73 o/o). 1H NMR (300 MHz, dβDMSO) δ 8.51 (s, I H), 7.94 (d, J = 9.1 Hz, 1 H), 7.73 (d, J = 8.9 Hz, 1 H), 4.14 (s, 3H), 2.67 (s, 3H). MS (ES+, m/z) 192 (M+H).

To a stirred solution of 2,3-dimethyl-6-nitro-2V-indazole (1.13 g) in 2- methoxyethyl ether (12 ml), at 0 °C, was added a solution of 4.48 g of tin(ll) chloride in 8.9 ml of concentrated HCI dropwise over 5 min. After the addition was complete, the ice bath was removed and the solution was allowed to stir for an additional 30 min. Approximately 40 ml of diethyl ether was added to reaction, resulting in precipitate formation. The resulting precipitate was isolated by filtration and washed with diethyl ether, and afforded a yellow solid (1.1 g, 95 %), the HCI salt 2,3-dimethyl-2/7-indazol-6- amine. 1H NMR (300 MHz, deDMSO) δ 7.77 (d, J = 8.9 Hz, 1 H), 7.18 (s, 1 H), 7.88 (m, 1 H), 4.04 (s, 3H), 2.61 (s, 3H). MS (ES+, m/z) 162 (M+H).

CLIP

Download PDF – Innovare Academic Sciences

innovareacademics.in/journals/index.php/ijpps/article/download/10558/4196
NMR, MASS

CLIP FROM

Let’s Research !!!!!

https://ayurajan.blogspot.in/2014/12/pazopanib.html

WO2003106416A2 (same appears in Drug Future 2006, 31, 7, 585-589)
Pazopanib synthesis: J Med Chem 2008, 51, 4632-4640 (same appears in Beilstein J Org Chem 2011, 7, 442–495)

CLIP

LINK

STR1 STR2 STR3

PAPER

http://www.eurekaselect.com/97375

10.2174/157017812800233714

A Novel Practical Synthesis of Pazopanib: An Anticancer Drug

Author(s): YiCheng Mei, BaoWei Yang, Wei Chen, DanDan Huang, Ying Li, Xin Deng, BaoMing Liu, JingJie Wang, Hai Qian and WenLong Huang

Affiliation: Center of Drug Discovery, China Pharmaceutical University, 24 Tongjiaxiang, Nanjing, Jiangsu 210009, P.R. China.

Abstract:

This paper reports a novel approach to synthesize pazopanib. In our synthetic route, the potently mutagenic alkylating agents such as dimethyl sulfate and methyl iodide are avoided. A novel regioselective methylation of the 2- position of 3-methyl-6-nitro-1H-indazole was reported. This novel route is one step shorter than the previously reported route.

PATENT

https://www.google.com/patents/WO2014097152A1?cl=en

Pazopanib is a tyrosine kinase inhibitor of Formula la.

Figure imgf000002_0001

Formula la

Pazopanib is marketed as the hydrochloride salt, with the chemical name 5-[[4- [(2,3-dimethyl-2H-indazol-6-yl)methylamino]-2-pyrimidinyl]amino]-2- methylbenzenesulfonamide monohydrochloride, having the structure as depicted in Formula I:

Figure imgf000002_0002

Formula I

U.S. Patent No. 7,105,530 provides a process for the preparation of a hydrochloride salt of a compound of Formula II

Figure imgf000003_0001

Formula II involving the reduction of 2,3-dimethyl-6-nitro-2H-indazole with tin (II) chloride in concentrated hydrochloric acid in the presence of 2-methoxyethyl ether at 0°C. It also describes the preparation of a compound of Formula III

Figure imgf000003_0002

Formula III involving the reaction of a hydrochloride salt of compound of Formula II with 2,4- dichloropyrimidine in the presence of a base and solvent mixture of

tetrahydrofuran/ethanol followed by stirring for 4 hours at 85°C.

PCT Publication No. WO 2007/064752 provides a process for the preparation of a compound of Formula II comprising reducing 2,3-dimethyl-6-nitro-2H-indazole with 10% Palladium-carbon (50% wet) in the presence of methanol, followed by the addition of ammonium formate at a rate that ensures the reaction temperature is maintained at or between 25°C and 30°C. It also discloses the preparation of a compound of Formula III comprising heating the compound of Formula II with sodium bicarbonate in presence of tetrahydrofuran and ethanol at or between 75°C and 80°C followed by cooling to 20°C to

25°C.

The present invention provides a process for the preparation of a compound of Formula II which offers recycling of the Raney nickel catalyst used in the process, and an easy filtration work-up procedure. Further, the present invention offers selective reduction under mild conditions that is economical to use at an industrial scale.

The present invention also provides a process for the preparation of compound of Formula III which avoids the use of two or more solvents, and additionally, also circumvents heating and cooling procedures during the reaction. The aforesaid advantages yield a compound of Formula III with a lesser amount of N-(4-chloropyrimidin-2-yl)-2,3- dimethyl-2H-indazol-6-amine (CPDMI) impurity.

The compounds of Formula II and Formula III prepared by the present invention yield a compound of Formula la or its salts in comparable yield and suitable purity required for medicinal preparations.

EXAMPLES

Step 1: Synthesis of 2,3-dimcthyl-6-nitro-2H-indazole

Example 1 :

Trimethyloxonium tetrafluoroborate (125.2 g, 0.85 mol) was added to a stirred suspension of 3-methyl-6-nitro-indazole (100 g, 0.56 mol) in ethyl acetate (2000 mL) over a period of 4 hours in four equal lots at 1 hour time intervals. The reaction mixture was stirred at 25 °C to 30°C for 16 hours. The solvent was recovered under reduced pressure. A saturated sodium bicarbonate solution (3240 mL) was added to the mixture slowly, and the reaction mixture was extracted with 4: 1 mixture of dichloromethane:isopropyl alcohol (1080 mL x 5). The solvent was recovered under reduced pressure. Methyl fert-butyl ether (800 mL) was added to the residue, and the reaction mixture was stirred for 30 minutes at 45 °C to 50°C. The reaction mixture was cooled to 25 °C to 30°C and was stirred at this temperature for 30 minutes. The solid was filtered, washed with methyl tert- butyl ether (100 mL x 2), and dried in an air oven at 50°C for 12 hours to afford 2,3- dimethyl-6-nitro-2H-indazole as a yellow solid.

Yield: 82.4% w/w

Step 2: Synthesis of 2,3-dimethyl-2H-indazol-6-amine

Example 2a:

Raney nickel ( 12.50 g) was added to a suspension of 2,3-dimethyl-6-nitro-2H- indazole (50 g, 0.26 mol) in methanol (500 mL). The reaction mixture was stirred in an autoclave under hydrogen pressure of 3.5 kg/cm2 – 4.0 kg/cm2 at 25°C to 30°C for 5 hours. Further, the reaction mixture was filtered through a hyflo bed, and the catalyst was washed with methanol (100 mL x 2). The filtrates were combined, and the solvent was recovered completely. «-Heptane (250 mL) and dichloromethane (50 mL) were added to the residue, and the reaction mixture was stirred for 1 hour at 25°C to 30°C. The solid was collected by filtration, washed with n-heptane (50 mL x 2), and dried under vacuum at 40°C to 45 °C to afford 2,3-dimethyl-2H-indazol-6-amine as a light brown solid.

Yield: 95% w/w

Example 2b:

Raney nickel (21.25 g) was added to a suspension of 2,3-dimethyl-6-nitro-2H- indazole (85 g, 0.45 mol) in methanol (850 mL). The reaction mixture was stirred in an autoclave under hydrogen pressure of 3.5 kg/cm2 – 4.0 kg/cm2 at 25°C to 30°C for 5 hours. Further, the reaction mixture was filtered through a hyflo bed, and the catalyst was washed with methanol (85 mL x 3). The filtrates were combined, and the solvent was recovered up to the volume of 850 mL. The 2,3-dimethyl-2H-indazol-6-amine in methanol was used as such in the next step. Step 3: Synthesis of N-(2-chloropyrimidin-4-yl)-2,3-dimethyl-2H-indazol-6-amine

Example 3 :

Sodium bicarbonate ( 112 g, 1.34 mol) was added to a stirred solution of 2,3- dimethyl-2H-indazol-6-amine (as obtained from step 2; Examples 2a and 2b) in methanol. 2,4-Dichloropyrimidine (99.35 g, 0.67 mol) was added to the reaction mixture followed by stirring of the reaction mixture for 24 hours at 25°C to 30°C. De-ionized water (850 mL) was added to the reaction mixture followed by stirring of the reaction mixture at 25 °C to 30°C for 1 hour. The solid was filtered. The wet solid was washed with de-ionized water (170 mL x 2) to obtain a wet material. De-ionized water (850 mL) was added to the wet material to obtain a slurry, and the slurry was stirred at 25°C to 30°C for 30 minutes. The solid was filtered, then washed with de-ionized water (170 mL x 2). The wet material obtained was treated with ethyl acetate (340 mL) to obtain a slurry. The slurry was stirred at 35°C to 40°C for 30 minutes and then cooled to 0°C to 5°C. The slurry was further stirred at 0°C to 5°C for 30 minutes. The solid was collected by filtration, then washed with cold ethyl acetate (170 mL x 2). The solid was dried in an air oven at 50°C for 16 hours to afford N-(2-chloropyrimidin-4-yl)-2,3 -dimethyl -2H-indazol-6-amine as an off- white solid.

Yield: 86.7% w/w

Step 4: Synthesis of pazopanib hydrochloride

Example 4a: Synthesis of N-(2-Chloropyrimidin-4-yl)-N.2.3-trimethyl-2H-indazol-6- amine

Cesium carbonate (238 g, 0.73 mol) and iodomethane (57 g, 0.40 mol) were added to a stirred suspension of N-(2-chloropyrimidin-4-yl)-2,3-dimethyl-2H-indazol-6-amine (lOOg, 0.37 mol) in N,N-dimethylformamide (300 mL) at 25°C to 30°C. The reaction mixture was further stirred at 25 °C to 30°C for 6 hours followed by cooling of the reaction mixture to 0°C to 5°C. De-ionized water (300 mL) was added drop-wise to the reaction mixture, then the reaction mixture was stirred at 5°C to 10°C for 30 minutes. The solid was collected by filtration, and washed with de-ionized water (100 mL x 2). The wet material so obtained was dried in an air oven at 50°C for 12 hours to obtain the title compound.

Yield: 90.4% w/w Example 4b: Synthesis of pazopanib hydrochloride

To a suspension of N-(2-chloropyrimidin-4-yl)-N-2,3-trimethyl-2H-indazol-6- amine (90 g, 0.312 mol) and 5-amino-2-methyl benzene sulfonamide (64.07 g, 0.344 mol) in isopropyl alcohol (900 mL) was added 4M hydrochloric acid solution in isopropyl alcohol (1.56 mL, 6.25 mol). The reaction mixture was heated to reflux temperature for 10 hours to 12 hours. The reaction mixture was cooled to 25°C. The reaction mixture was further stirred at 25°C to 30°C for 30 minutes, then the solid was filtered. The wet solid was washed with isopropyl alcohol (180 mL x 2), and then dried under vacuum at 45 °C to 50°C for 12 hours to afford the hydrochloride salt of 5-({4-[(2,3-dimethyl-21-I-indazol-6- yl)(methyl) amino] pyrimidin-2-yl} amino-Z-methylbenzene sulfonamide as a light brown solid.

Yield: 97% w/w

PATENT

https://www.google.com/patents/CN104557881A?cl=en

pazopanib hydrochloride monohydrate prepared:

(1) chemical reaction formula

Figure CN104557881AD00051

(2) Operation process

In the reaction flask pazopanib hydrochloride crude 100g, 700ml of acetonitrile was added under stirring and purified water 200ml, feeding is completed, begin heating to 75~80 ° C, until clear solvent filtration system, slowly dropped 10~20 ° C, keep stirring lh, filtered, and the filter cake washed with purified water and acetonitrile respectively, drained and the filter cake was dried at 60 ° C blast 5h, have pazopanib hydrochloride monohydrate solid 84g, yield 80.9%. For example, crystalline form pazopanib hydrochloride prepared the following examples.

pazopanib hydrochloride polymorph of preparation:

Example 1:

 In the reaction flask pazopanib hydrochloride monohydrate 8. 0g, ethanol 50ml and purified water 0.Iml (the volume of water accounted for a mixed solution of 2% of the total volume of the square, ethanol – water mixture total volume was 6.26 times pazopanib hydrochloride monohydrate quality), heated to 75 ° C, stirred at reflux for about 5h, after cooling to 10~20 ° C, keep stirring lh, the filter cake washed with ethanol, then blast drying at 105 ° C 5h, to obtain ultrafine powder solid 6. 8g, yield of 81. 9%, HPLC purity was 99.8%, as measured crystal X- ray powder diffraction pattern of FIG. 1 the basic consistent, as measured with a DSC thermogram consistent FIG. 2, the particle size distribution measurement is basically the same as Fig 3 (D90 <10ym).

CLIP

Pazopanib is a highly bio-available, multi- tyrosine kinase inhibitor of vascular endothelial growth factor receptor (VEGFR)-l, -2, -3, platelet-derived factor receptor (PDGFR) -α, -β, cytokine receptor (cKit), interleukin-2 receptor inducible T-cell kinase (Itk), leukocyte-specific protein tyrosine kinase (Lck), and transmembrane glycoprotein receptor tyrosine kinase (c-Fms). Pazopanib was recently approved by the Food and Drug Administration (FDA) for the treatment of patients with advanced renal cell carcinoma; thus adding to the other FDA-approved VEGF pathway inhibitors, sunitinib, bevacizumab (in combination with interferon) and sorafinib for this same indication.

Processes by which pazopanib and its intermediates can be synthesized have been described in US Patent No. 7,105,530 as well as in the published PCT application WO03/106416.

U.S. patent no. 7,105,530 disclosed pyrimidineamines and their derivatives thereof. These compounds are antineoplastic agents, and are useful in the treatment of various cancers and renal cell carcinoma. Among them pazopanib hydrochloride, chemically 5-[4-[N-(2,3-Dimethyl-2H-indazol-6-yl)-N-methylamino]pyrimidin-2- ylamino]-2-methylbenzenesulfonamide hydrochloride. Pazopanib hydrochloride is represented by the following structure:
Pazopanib hydrochloride is a potent and selective multi-targeted receptor tyrosine kinase inhibitor of VEGFR (Vascular endothelial growth factor receptors)- 1, VEGFR-2, VEGFR-3, PDGFR (Platelet-derived growth factor receptors )-a/p, and c-kit that blocks tumor growth and inhibits angiogenesis. It has been approved for renal cell carcinoma by the U.S. Food and Drug Administration. Pazopanib hydrochloride may also be active in ovarian cancer and soft tissue sarcoma. Pazopanib hydrochloride also appears effective in the treatment of non-small cell lung carcinoma. Pazopanib hydrochloride is marketed under the brand name Votrient® by Glaxosmithkline in the form of tablet.

Processes for the preparation of pazopanib hydrochloride and related compounds were disclosed in U.S. patent no. 7,105,530 and U.S. patent no. 7,262,203.

According to U.S. patent no. 7,105,530, pazopanib hydrochloride can be prepared by reacting the N-(2-chloropyrimidin-4-yl)-N,2,3-trimethyl-2H-indazol-6-amine with 5- amino-2-methylbenzenesulfonamide in the presence of hydrochloric acid in isopropanol and ether.

U.S. patent application publication no. 2006/0252943 disclosed a process for the preparation of pazopanib hydrochloride. According to this patent, pazopanib hydrochloride can be prepared by reacting the N-(2-chloropyrimidin-4-yl)-N,2,3- trimethyl-2H-indazol-6-amine with 5-amino-2-methylbenzenesulfonamide in the presence of hydrochloric acid in ethanol or methanol or tetrahydrofuran or acetonitrile and dioxane.

Drugs of the Future, 2006, 31 (7): 585-589
WO0306416

CLIP

Marcus BaumannEmail of corresponding author, Ian R. BaxendaleEmail of corresponding author, Steven V. LeyEmail of corresponding author and Nikzad NikbinEmail of corresponding author
Innovative Technology Centre, Department of Chemistry, University of Cambridge, Lensfield Road, CB2 1EW Cambridge, UK
Email of corresponding author Corresponding author email
Editor-in-Chief: J. Clayden
Beilstein J. Org. Chem. 2011, 7, 442–495.

Pazopanib (246, Votrient) is a new potent multi-target tyrosine kinase inhibitor for various human cancer cell lines. Pazopanib is considered a promising replacement treatment to imatinib and sunitinib and was approved for renal cell carcinoma by the FDA in late 2009. The indazole system is built up via diazotisation and spontaneous cyclisation of 2-ethyl-5-nitroaniline (247) using tert-butyl nitrite. The resulting indazole structure 249 can be methylated entirely regioselectively with either Meerwein’s salt, trimethyl orthoformate or dimethyl sulfate. A tin-mediated reduction of the nitro group unmasks the aniline which undergoes nucleophilic aromatic substitution to introduce the pyrimidine system with the formation of 253. Methylation of the secondary amine function with methyl iodide prior to a second SNAr reaction with a sulfonamide-derived aniline affords pazopanib .

[1860-5397-7-57-i50]
Synthesis of pazopanib.
  1. Pandite, A. N.; Whitehead, B. F.; Ho, P. T. C.; Suttle, A. B. Cancer Treatment Method. WO Patent 2007/064753, June 7, 2007.
  2. Harris, P. A.; Boloor, A.; Cheung, M.; Kumar, R.; Crosby, R. M.; Davis-Ward, R. G.; Epperly, A. H.; Hinkle, K. W.; Hunter, R. N., III; Johnson, J. H.; Knick, V. B.; Laudeman, C. P.; Luttrell, D. K.; Mook, R. A.; Nolte, R. T.; Rudolph, S. K.; Szewczyk, J. R.; Truesdale, A. T.; Veal, J. M.; Wang, L.; Stafford, J. A. J. Med. Chem.2008,51,4632–4640. doi:10.1021/jm800566m

CLIP

STR1

Pazopanib hydrochloride (Votrient)
Pazopanib is a potent and selective multi-targeted receptor tyrosine kinase inhibitor of VEGFR-1, VEGFR-2, VEGFR-3, PDGFR-a/b, and c-kit that blocks tumor growth and inhibits angiogenesis. It was approved for renal cell carcinoma by the U.S. Food
and Drug Administration in 2009 and is marketed under the trade name Votrient by the drug’s manufacturer, GlaxoSmithKline. The
synthesis of pazopanib begins with methylation of 3-methyl-6-nitroindazole (82) with trimethyl orthoformate in the presence of BF3OEt to give indazole 83 in 65% yield (Scheme 14).65 Reduction of the nitro group was achieved via transfer hydrogenation to give 84 in 97% yield, and this was followed by coupling the aniline with 2,4-dichloropyrimidine in a THF-ethanol mixture at elevated
temperature to provide diarylamine 85 in 90% yield. The aniline nitrogen was then methylated using methyl iodide to give 86 in
83% yield prior to coupling with 5-amino-2-methylbenzenesulfonamide (87) and salt formation using an alcoholic solution of
HCl to furnish pazopanib hydrochloride (XIV) in 81% yield.

STR4

FDA Orange Book Patents

FDA Orange Book Patents: 1 of 3
Patent 7262203
Expiration Dec 19, 2021
Applicant NOVARTIS PHARMS CORP
Drug Application
  1. N022465 (Discontinued Drug: VOTRIENT. Ingredients: PAZOPANIB HYDROCHLORIDE)
  2. N022465 (Prescription Drug: VOTRIENT. Ingredients: PAZOPANIB HYDROCHLORIDE)
 
FDA Orange Book Patents: 2 of 3
Patent 8114885
Expiration Dec 19, 2021
Applicant NOVARTIS PHARMS CORP
Drug Application
  1. N022465 (Discontinued Drug: VOTRIENT. Ingredients: PAZOPANIB HYDROCHLORIDE)
  2. N022465 (Prescription Drug: VOTRIENT. Ingredients: PAZOPANIB HYDROCHLORIDE)
FDA Orange Book Patents: 3 of 3
Patent 7105530
Expiration Oct 19, 2023
Applicant NOVARTIS PHARMS CORP
Drug Application
  1. N022465 (Discontinued Drug: VOTRIENT. Ingredients: PAZOPANIB HYDROCHLORIDE)
  2. N022465 (Prescription Drug: VOTRIENT. Ingredients: PAZOPANIB HYDROCHLORIDE)

VOTRIENT (pazopanib) is a tyrosine kinase inhibitor (TKI). Pazopanib is presented as the hydrochloride salt, with the chemical name 5-[[4-[(2,3-dimethyl-2H-indazol-6- yl)methylamino]-2-pyrimidinyl]amino]-2-methylbenzenesulfonamide monohydrochloride. It has the molecular formula C21H23N7O2S•HCl and a molecular weight of 473.99. Pazopanib hydrochloride has the following chemical structure:

VOTRIENT (pazopanib) Structural Formula Illustration

Pazopanib hydrochloride is a white to slightly yellow solid. It is very slightly soluble at pH 1 and practically insoluble above pH 4 in aqueous media.

Tablets of VOTRIENT are for oral administration. Each 200 mg tablet of VOTRIENT contains 216.7 mg of pazopanib hydrochloride, equivalent to 200 mg of pazopanib free base. The inactive ingredients of VOTRIENT are:Tablet Core: Magnesium stearate, microcrystalline cellulose, povidone, sodium starch glycolate. Coating: Gray film-coat: Hypromellose, iron oxide black, macrogol/polyethylene glycol 400 (PEG 400), polysorbate 80, titanium dioxide.

  1. FierceBiotech. 2008-09-15. Retrieved 2010-08-10.
Country
Patent Number
Approved
Expires (estimated)
United States 7105530 2009-10-19 2023-10-19
United States 7262203 2009-10-19 2021-12-19
United States 8114885 2009-10-19 2021-12-19

JUNE 4 2013 old article cut paste

GlaxoSmithKline’s (GSK) Votrient (pazopanib) has met the primary objective of a statistically significant improvement in the time to disease progression or death that is the progression-free survival (PFS) against placebo in Phase III ovarian cancer..

http://clinicaltrials.pharmaceutical-business-review.com/news/gsks-votrient-meets-primary-objective-in-phase-iii-ovarian-cancer-trial-030613

Pazopanib shrinks lung cancers before surgery

FORMULATION

https://www.google.co.in/patents/US20140255505

Pazopanib is an angiogenesis inhibitor targeting vascular endothelial growth factor receptors (VEGFR)-1, -2, and -3, platelet-derived growth factor receptors (PDGFR)-α/-β, and c-Kit. The hydrochloride salt of pazopanib (5-[[4-[(2,3-dimethyl-2H-indazol-6-yl)methylamino]-2-pyrimidinyl]amino]-2-methylbenzenesulfonamide) is marketed by GlaxoSmithKline as Votrient®, which is approved in the United States and other countries for the treatment of renal cell carcinoma (RCC).

Votrient® is currently prescribed to adults in the form of 200 mg tablets for oral administration, with each 200 mg tablet containing an amount of pazopanib hydrochloride equivalent to 200 mg of pazopanib free base.

Though the current tablets are acceptable of use in adults, the tablets are not preferred for use in potential future use for administering pazopanib to children. In pediatric populations, it is often desired that drug be available as a powder for reconstitution to an oral suspension. Manufacture of such a powder requires dry blending of various excipients with the active substance to provide good flow properties and content uniformity of the powder blend.

Several additional challenges exist concerning the use of pazopanib in a pediatric formulation. For instance, the nature of the drug substance favors conversion from the hydrochloride salt to the free base and hydrate forms in an aqueous environment such that standard formulations fail to provide adequate suspension stability at long term storage conditions of 25° C./65% RH or room temperature. Further, the drug has been found to have a bitter taste and, therefore, taste masking is critical.It is desired to invent a pediatric formulation of pazopanib hydrochloride suitable for administration to a pediatric population

References

  1.  “Votrient (pazopanib) dosing, indications, interactions, adverse effects, and more”. Medscape Reference. WebMD. Retrieved 27 January 2014.
  2.  “VOTRIENT (pazopanib hydrochloride) tablet, film coated [GlaxoSmithKline LLC]”(PDF). DailyMed. GlaxoSmithKline LLC. November 2013. Retrieved 27 January 2014.
  3.  “Votrient : EPAR – Product Information” (PDF). European Medicines Agency. Glaxo Group Ltd. 23 January 2014. Retrieved 27 January 2014.
  4.  “Votrient 200 mg and 400 mg film coated tablets – Summary of Product Characteristics (SPC)”. electronic Medicines Compendium. GlaxoSmithKline UK. 20 December 2013. Retrieved 27 January 2014.
  5.  “PRODUCT INFORMATION VOTRIENT® TABLETS” (PDF). TGA eBusiness Services. GlaxoSmithKline Australia Pty Ltd. 25 March 2013. Retrieved 27 January 2014.
  6.  “Pharmaceutical Benefits Scheme (PBS) – Pazopanib”. Pharmaceutical Benefits Scheme. Australian Government. Retrieved 27 January 2014.
  7.  “Pazopanib – Online Pharmaceutical Schedule”. Pharmaceutical Management Agency. Retrieved 9 June 2015.
  8. ^ “Pazopanib shows encouraging activity in several tumour types, including soft tissue sarcoma and ovarian cancer”. FierceBiotech. 2008-09-15. Retrieved 2010-08-10.
  9.  “GSK pulls bid to extend use of kidney drug to ovarian cancer”. Reuters. 31 March 2014. Retrieved 7 April 2014.
  10.  “Regulatory update: Votrient (pazopanib) as maintenance therapy for advanced ovarian cancer in the EU”. GlaxoSmithKline. 31 March 2014. Retrieved 7 April 2014.
  11. Zivi, A; Cerbone, L; Recine, F; Sternberg, CN (September 2012). “Safety and tolerability of pazopanib in the treatment of renal cell carcinoma”. Expert Opinion on Drug Safety. 11 (5): 851–859. doi:10.1517/14740338.2012.712108. PMID 22861374.
  12. Khurana V, Minocha M, Pal D, Mitra AK (March 2014). “Role of OATP-1B1 and/or OATP-1B3 in hepatic disposition of tyrosine kinase inhibitors.”. Drug Metabol Drug Interact. 0 (0): 1–11. doi:10.1515/dmdi-2013-0062. PMID 24643910.
  13.  Khurana V, Minocha M, Pal D, Mitra AK (May 2014). “Inhibition of OATP-1B1 and OATP-1B3 by tyrosine kinase inhibitors.”. Drug Metabol Drug Interact. 0 (0): 1–11.doi:10.1515/dmdi-2014-0014. PMID 24807167.
  14.  Verweij, J; Sleijfer, S (May 2013). “Pazopanib, a new therapy for metastatic soft tissue sarcoma”. Expert Opinion on Pharmacotherapy. 14 (7): 929–935.doi:10.1517/14656566.2013.780030. PMID 23488774.
  15.  Schöffski, P (June 2012). “Pazopanib in the treatment of soft tissue sarcoma”. Expert Review of Anticancer Therapy. 12 (6): 711–723. doi:10.1586/era.12.41.PMID 22716487.
  16.  Pick, AM; Nystrom, KK (March 2012). “Pazopanib for the treatment of metastatic renal cell carcinoma”. Clinical Therapeutics. 34 (3): 511–520.doi:10.1016/j.clinthera.2012.01.014. PMID 22341567.
  17. Rimel, BJ (April 2015). “Antiangiogenesis agents in ovarian cancer”. Contemporary Oncology. 7 (2): 16–19. PMID 21638926.
WO2003106416A2 * Jun 17, 2003 Dec 24, 2003 Smithkline Beecham Corporation Chemical process
WO2005054212A2 * Nov 26, 2004 Jun 16, 2005 Ciba Specialty Chemicals Holding Inc. Electroluminescent device
WO2007064752A2 Nov 29, 2006 Jun 7, 2007 Smithkline Beecham Corporation Treatment of ocular neovascular disorders such as macular degeneration, angiod streaks, uveitis and macular edema
WO2011069053A1 Dec 3, 2010 Jun 9, 2011 Teva Pharmaceutical Industries Ltd. Process for the preparation of pazopanip hcl and crystalline forms of pazopanib hcl
WO2012051659A1 * Oct 20, 2011 Apr 26, 2012 Biota Scientific Management Pty Ltd Viral polymerase inhibitors
US7105530 Dec 19, 2001 Sep 12, 2006 Smithkline Beecham Corporation Pyrimidineamines as angiogenesis modulators
Reference
1 * DAVIES R R: “Indazole derivatives: the synthesis of various amino- and hydroxy-indazoles and derived sulphonic acids“, JOURNAL OF THE CHEMICAL SOCIETY, CHEMICAL SOCIETY, LETCHWORTH; GB, 1 January 1955 (1955-01-01), pages 2412-2423, XP009176650, ISSN: 0368-1769, DOI: 10.1039/JR9550002412
US20060252943 * Jun 17, 2003 Nov 9, 2006 Amogh Boloor Chemical process
US20080269170 * Jan 9, 2008 Oct 30, 2008 Sanofi-Aventis Novel 2,4-Dianilinopyrimidine Derivatives, the Preparation Thereof, Their Use as Medicaments, Pharmaceutical Compositions and, in Particular, as IKK Inhibitors
Reference
1 * LUO G. ET AL.: “Microwave-assisted synthesis of aminopyrimidines“, TETRAHEDRON LETTERS, vol. 43, no. 33, 2002, pages 5739 – 5742, XP004372432
2 * See also references of EP2646431A4
Citing Patent Filing date Publication date Applicant Title
WO2014085373A1 * Nov 26, 2013 Jun 5, 2014 Glaxosmithkline Llc Combination
CN103232443A * Feb 1, 2013 Aug 7, 2013 天津药物研究院 Indazole derivative crystal and its preparation method and use
CN104557881A * Dec 30, 2014 Apr 29, 2015 山东博迈康药物研究有限公司 Preparation method of pazopanib hydrochloride crystal form
Boloor, A.; et. al. Pyrimidineamines as angiogenesis modulators. US7105530B2
2. Boloor, A.; Harris, P. A.; et. al. Discovery of 5-[[4-[(2,3-Dimethyl-2H-indazol-6-yl)methylamino]-2-pyrimidinyl]amino]-2-methyl-benzenesulfonamide (Pazopanib), a Novel and Potent Vascular Endothelial Growth Factor Receptor Inhibitor. J Med Chem 2008, 51(15), 4632–4640.
3. Bhanushali, D. S.; et. al. Compositions and processes. WO2011050159A1
4. Boloor, A.; et. al.  Chemical Process. WO2003106416A2
5. Pandite, A. M.; et. al. Cancer treatment method. WO2007064753A2
European Medicines Agency, “CHMP assessment report: Votrient“, pages 1, 5, 6 (2010).

Click to access 022465s000_ChemR.pdf

Pazopanib
Pazopanib.svg
Pazopanib3Dan.gif
Systematic (IUPAC) name
5-[[4-[(2,3-Dimethyl-2H-indazol-6-yl)methylamino]-2-pyrimidinyl]amino]-2-methylbenzolsulfonamide
Clinical data
Trade names Votrient
AHFS/Drugs.com Monograph
MedlinePlus a610013
License data
Pregnancy
category
  • AU: D
  • US: D (Evidence of risk)
Routes of
administration
Oral
Legal status
Legal status
Pharmacokinetic data
Protein binding >99%[1]
Metabolism Hepatic (CYP3A4, 1A2 and2C8-mediated)[1]
Biological half-life 31.9 hours[1]
Excretion Faeces (primary), urine (<4%)[1]
Identifiers
CAS Number 444731-52-6 
ATC code L01XE11 (WHO)
PubChem CID 11525740
ChemSpider 9700526 Yes
UNII 7RN5DR86CK Yes
ChEMBL CHEMBL477772 
Chemical data
Formula C21H23N7O2S
Molar mass 437.517 g/mol

////////////PAZOPANIB, GW786034, Votrient, Armala, GW 786034, GW-786034, GW786034GW786034, VOTRIENT, Pazopanib hydrochloride, FDA 2009, Antineoplastic,  Tyrosine Kinase Inhibitors, Protein Kinase Inhibitors,  Renal Cell Carcinoma Therpay,  Soft Tissue Sarcoma Therapy, パゾパニブ塩酸塩 , Пазопаниба Гидрохлорид

O=S(=O)(N)c1c(ccc(c1)Nc2nccc(n2)N(c4ccc3c(nn(c3C)C)c4)C)C