Home » Antineoplastic (Page 9)
Category Archives: Antineoplastic
 DRUG APPROVALS BY DR ANTHONY MELVIN CRASTO
.....FOR BLOG HOME CLICK HERE
Flag and hits
ORGANIC SPECTROSCOPY

SUBSCRIBE

 Subscribe in a reader
Enter your email address:
Delivered by FeedBurner
Subscribe to New Drug Approvals by Email


Nefextinib
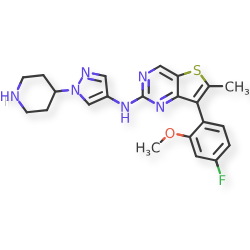
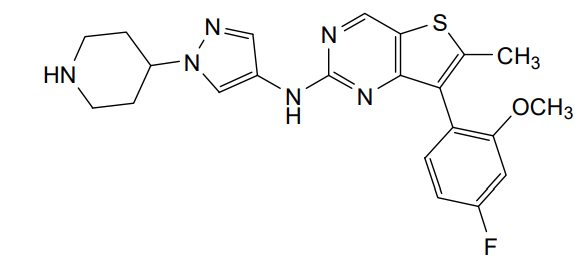
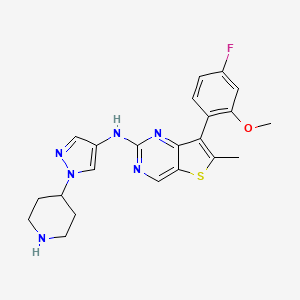
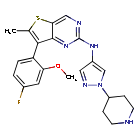
Nefextinib
CAS 2070931-57-4
MF C22H23FN6OS MW 438.52
7-(4-fluoro-2-methoxyphenyl)-6-methyl-N-[1-(piperidin4-yl)-1H-pyrazol-4-yl]thieno[3,2-d]pyrimidin-2-amine
7-(4-FLUORO-2-METHOXYPHENYL)-6-METHYL-N-(1-(PIPERIDIN-4-YL)-1H-PYRAZOL-4-YL) THIENO (3,2-D)PYRIMIDIN-2-AMINE
tyrosine kinase inhibitor, antineoplastic, DL772G3NN7, MAX-40279, MAX 40279
Nefextinib is an orally bioavailable inhibitor of the fibroblast growth factor receptor (FGFR) and FMS-like tyrosine kinase 3 (FLT3; CD135; STK1; FLK2), with potential antineoplastic activity. Upon oral administration, nefextinib binds to and inhibits both FGFR and FLT3, including FLT3 mutant forms, which results in the inhibition of FGFR/FLT3-mediated signal transduction pathways. This inhibits proliferation in FGFR/FLT3-overexpressing tumor cells. FGFR, a family of receptor tyrosine kinases, is upregulated in many tumor cell types. FLT3, a class III receptor tyrosine kinase (RTK), is overexpressed or mutated in most B-lineage neoplasms and in acute myeloid leukemias. They both play key roles in cellular proliferation and survival.
- A Phase 2 Study to Evaluate the Safety and Efficacy of Max-40279-01 in Patients With Advanced Gastric Cancer or Gastroesophageal Junction CancerCTID: NCT05395780Phase: Phase 2Status: Unknown statusDate: 2022-06-02
- MAX-40279 in Subjects With Acute Myelogenous Leukemia (AML)CTID: NCT03412292Phase: Phase 1Status: Unknown statusDate: 2022-01-19
- MAX-40279-01 in Patients With Advanced Solid TumorsCTID: NCT04183764Phase: Phase 1Status: Unknown statusDate: 2022-01-19
- Study of MAX-40279 in Patients With Relapsed or Refractory Acute Myelogenous Leukemia (AML)CTID: NCT04187495Phase: Phase 1Status: Unknown statusDate: 2022-01-19
- A Clincal Study of Max-40279-01 in Patients With Advanced Colorectal CancerCTID: NCT05130021Phase: Phase 2Status: Unknown statusDate: 2021-12-06
SYN
SYN
https://patentscope.wipo.int/search/en/detail.jsf?docId=WO2017012559&_cid=P22-MHRG1L-67142-1
[0488]N-[7-(4-fluoro-2-methoxyphenyl)-6-methylthieno[3,2-d]pyrimidin-2-yl]-1-(piperidin-4-yl)-1H-pyrazol-4-amine (compound 31)
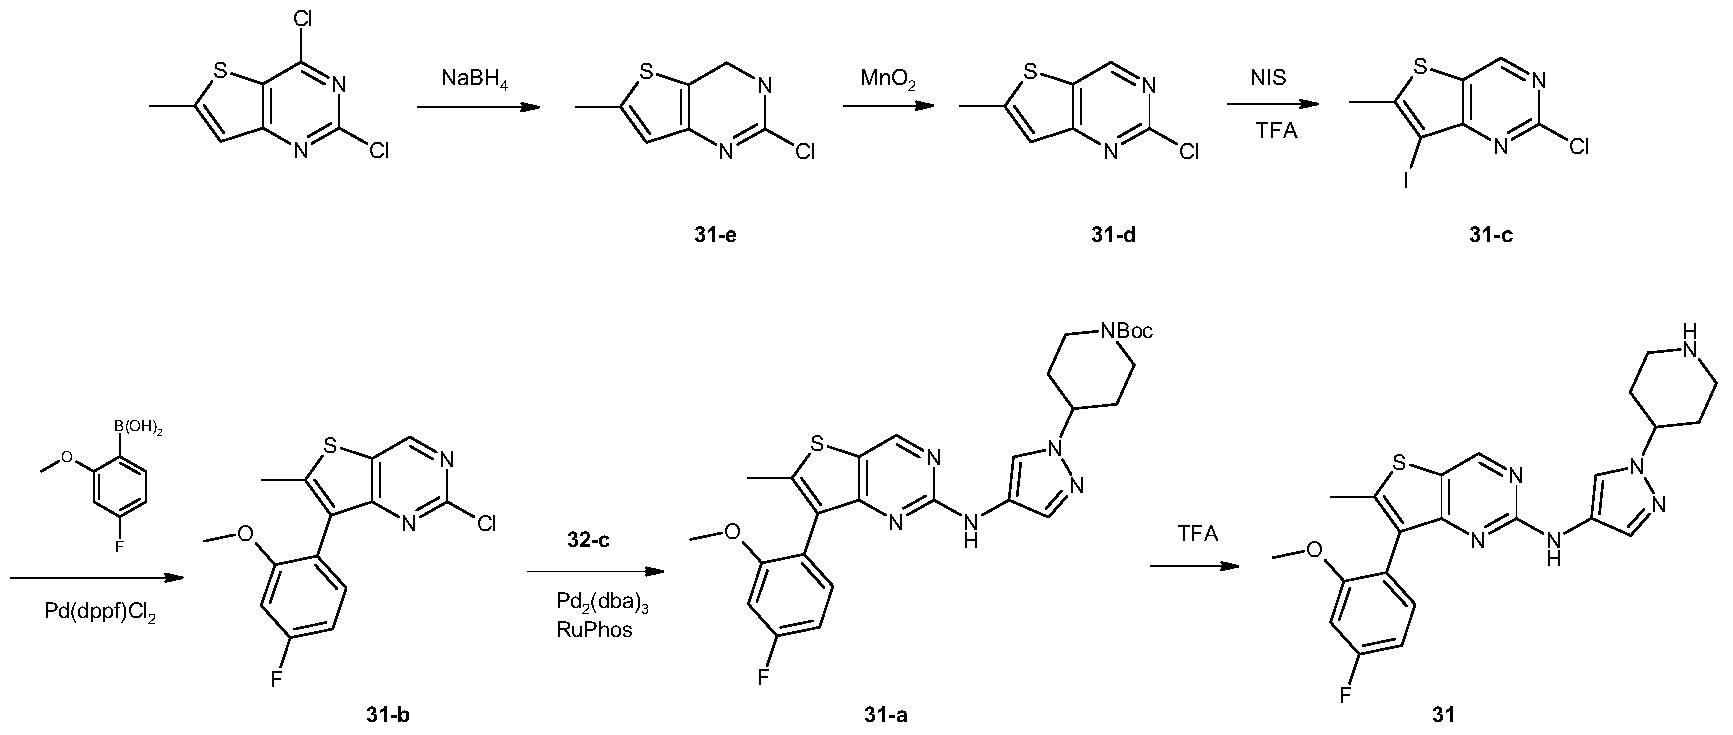
[0491]2,4-Dichloro-6-methylthiophene[3,2-d]pyrimidine (10 g, 45.6 mmol) was dissolved in tetrahydrofuran (100 mL) and ethanol (100 mL). The reaction mixture was cooled to 0 °C, and sodium borohydride (12.5 g, 198 mmol) was added in portions. The reaction mixture was brought to room temperature and stirred for 16 hours. It was then diluted with water (500 mL) and adjusted to pH 7 with 1 N hydrochloric acid solution. The aqueous phase was extracted with ethyl acetate (150 mL × 3). The organic phase was washed successively with water (100 mL × 3) and saturated brine (100 mL), dried over anhydrous sodium sulfate, filtered, and the filtrate was concentrated under reduced pressure to give a white solid 31-e (7.5 g, yield: 88%). This product required no further purification. LC-MS (ESI): m/z = 187 [M+H] + .
[0492]Synthesis of compound 31-d
[0493]Compound 31-e (7.5 g, 40 mmol) was dissolved in chloroform (300 mL) at 0 °C, and activated manganese dioxide (35 g, 400 mmol) was added. The reaction mixture was brought to room temperature and stirred for 16 hours. The reaction mixture was filtered through diatomaceous earth, and the filter cake was washed with chloroform (100 mL × 3). The combined filtrates were concentrated under reduced pressure to give a white solid 31-d (6.6 g, yield: 89%), which did not require further purification. LC-MS (ESI): m/z = 185 [M + H]+.
[0494]Synthesis of compound 31-c
[0495]Compound 31-d (3.1 g, 16.8 mmol) was dissolved in trifluoroacetic acid (30 mL) at 0 °C. N-iodosuccinimide (5.7 g, 25.3 mmol) was added in portions. The reaction mixture was brought to room temperature and stirred for 1 hour. The reaction was quenched with water (50 mL) and extracted with dichloromethane (50 mL × 3). The organic phase was washed successively with water (50 mL × 3) and saturated brine (50 mL), dried over anhydrous sodium sulfate, filtered, and the filtrate was concentrated under reduced pressure to give a white solid 31-c (4.9 g, yield: 94%). This product required no further purification. LC-MS (ESI): m/z = 311 [M + H] + .
[0496]Synthesis of compound 31-b
[0497]Compound 31-c (615 mg, 1.98 mmol), 2-methoxy-4-fluorophenylboronic acid (405 mg, 2.38 mmol), and sodium carbonate (630 mg, 5.94 mmol) were suspended in dioxane (5 mL) and water (5 mL). A [1,1′-bis(diphenylphosphine)ferrocene]palladium dichloride dichloromethane complex (163 mg, 0.2 mmol) was added. The mixture was purged three times with nitrogen and heated to 80 °C for 16 hours. After cooling to room temperature, the reaction solution was concentrated under reduced pressure. The residue was separated into layers by dichloromethane (50 mL) and water (50 mL). The organic phase was dried over anhydrous sodium sulfate, filtered, and the filtrate was concentrated and purified by silica gel column chromatography (petroleum ether:dichloromethane = 1:1) to give a white solid 31-b (240 mg, yield: 39%). LC-MS (ESI): m/z = 309 [M+H] + .
[0498]Synthesis of compound 31-a
[0499]Compound 31-b (240 mg, 0.78 mmol) and compound 32-c (208 mg, 0.78 mmol) were dissolved in N,N-dimethylformamide (3 mL), and potassium carbonate (323 mg, 2.34 mmol), 2-dicyclohexylphosphine-2′,6′-diisopropoxy-1,1′-biphenyl (112 mg, 0.24 mmol), and tris(dibenzylacetone)palladium (134 mg, 0.24 mmol) were added. The reaction was carried out under nitrogen protection at 110 °C for 16 hours. After cooling to room temperature, the reaction mixture was separated into layers by dichloromethane (50 mL) and water (50 mL). The organic phase was dried over anhydrous sodium sulfate, filtered, and the filtrate was concentrated under reduced pressure. The residue was purified by silica gel thin-layer chromatography (petroleum ether: ethyl acetate = 1:1) to give a yellow viscous oil 31-a (190 mg, yield: 45%). LC-MS(ESI): m/z = 539[M+H] + .
[0500]Synthesis of Compound 31
[0501]31-a (190 mg, 0.35 mmol) was dissolved in dichloromethane (3 mL), and trifluoroacetic acid (3 mL) was added. The mixture was stirred at room temperature for 3 hours. The reaction solution was concentrated under reduced pressure, and the residue was separated into layers by ethyl acetate (50 mL) and 1N hydrochloric acid aqueous solution (50 mL). The aqueous phase was adjusted to pH = 10 with saturated potassium carbonate aqueous solution, and a solid precipitated. The solid was filtered, and the filter cake was washed with water (20 mL × 3). The solid was dried under vacuum to give a light yellow solid 31 (22 mg, yield: 14%). LC-MS (ESI): m/z = 439 [M+H] + .
[0502]
1H-NMR(400MHz,MeOD)δ:8.78(d,J=5Hz,1H),7.87(s,1H),7.48(s,1H),7.35(m,1H),7.05(dd,J=11Hz,J=2Hz,1H),6.91(m,1H),4.10(m,1H),3.79(s,3H),3.22(m,2H),2.77(m,2H),2.47(s,3H),2.03(m,2H),1.73(m,2H)ppm
PAT
- Condensed ring pyrimidine compound, intermediate, its preparation method, composition and applicationPublication Number: CN-106366093-BPriority Date: 2015-07-21Grant Date: 2020-08-18
- Condensation ring pyrimidine compounds, intermediates, methods for producing them, compositions and applicationsPublication Number: JP-6875372-B2Priority Date: 2015-07-21Grant Date: 2021-05-26
- Condensed ring pyrimidine compounds, intermediates, preparation methods, compositions and applications thereofPublication Number: KR-102591886-B1Priority Date: 2015-07-21Grant Date: 2023-10-20
- Fused ring pyrimidine compound, intermediate, and preparation method, composition and use thereofPublication Number: EP-3354653-B1Priority Date: 2015-07-21Grant Date: 2019-09-04
- Fused ring pyrimidine compounds, intermediates, production methods, compositions and applications thereofPublication Number: JP-2018520202-APriority Date: 2015-07-21
- Fused ring pyrimidine compound, intermediate, and preparation method, composition and use thereofPublication Number: US-10494378-B2Priority Date: 2015-07-21Grant Date: 2019-12-03
- Fused ring pyrimidine compound, intermediate, and preparation method, composition and use thereofPublication Number: US-2018208604-A1Priority Date: 2015-07-21
- Fused ring pyrimidine compound, intermediate, and preparation method, composition and use thereofPublication Number: WO-2017012559-A1Priority Date: 2015-07-21


AS ON OCT2025 4.511 LAKHS VIEWS ON BLOG WORLDREACH AVAILABLEFOR YOUR ADVERTISEMENT
join me on Linkedin
Anthony Melvin Crasto Ph.D – India | LinkedIn
join me on Researchgate
RESEARCHGATE

join me on Facebook
Anthony Melvin Crasto Dr. | Facebook
join me on twitter
Anthony Melvin Crasto Dr. | twitter
+919321316780 call whatsaapp
EMAIL. amcrasto@gmail.com

……
//////////nefextinib, tyrosine kinase inhibitor, antineoplastic, DL772G3NN7, MAX-40279, MAX 40279
Lunbotinib
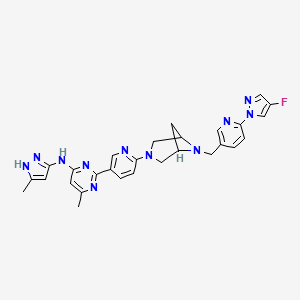
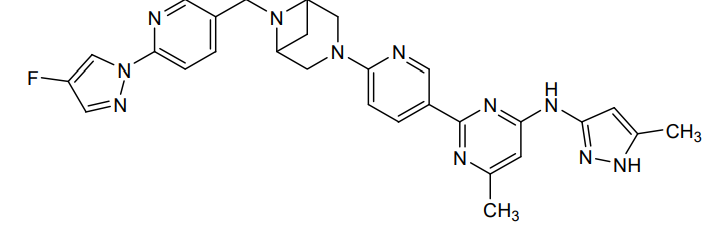
Lunbotinib
CAS 2479961-46-9
MF C28H28FN11 MW537.6 g/mol
2-[6-(6-{[6-(4-fluoro-1H-pyrazol-1-yl)pyridin-3-yl]methyl}-3,6-diazabicyclo[3.1.1]heptan-3-yl)pyridin-3-yl]-6-methyl-N-(5-methyl1H-pyrazol-3-yl)pyrimidin-4-amine
tyrosine kinase inhibitor, antineoplastic, KL3T9ZU6HQ
- 2-(6-(6-((6-(4-fluoropyrazol-1-yl)pyridin-3-yl)methyl)-3,6-diazabicyclo(3.1.1)heptan-3-yl)pyridin-3-yl)-6-methyl-N-(5-methyl-1H-pyrazol-3-yl)pyrimidin-4-amine
- 2-[6-[6-[[6-(4-fluoropyrazol-1-yl)pyridin-3-yl]methyl]-3,6-diazabicyclo[3.1.1]heptan-3-yl]pyridin-3-yl]-6-methyl-N-(5-methyl-1H-pyrazol-3-yl)pyrimidin-4-amine
Lunbotinib is an orally bioavailable selective inhibitor of the proto-oncogene receptor tyrosine kinase rearranged during transfection (RET), with potential antineoplastic activity. Upon oral administration, lunbotinib selectively binds to various RET fusions and mutations, including solvent front resistance mutations, and inhibits the activity of RET. This results in an inhibition of cell growth of tumors that exhibit increased RET activity due to these fusions and mutations. RET overexpression, activating mutations, and fusions result in the upregulation and/or overactivation of RET tyrosine kinase activity in various cancer cell types. Dysregulated RET activity plays a key role in the development and progression of certain cancers. Lunbotinib is able to penetrate the blood-brain barrier (BBB) and may also be able to overcome resistance mechanisms to first generation selective RET inhibitors (SRIs).
SYN
https://patentscope.wipo.int/search/en/detail.jsf?docId=WO2020168939&_cid=P12-MHKH7H-14851-1
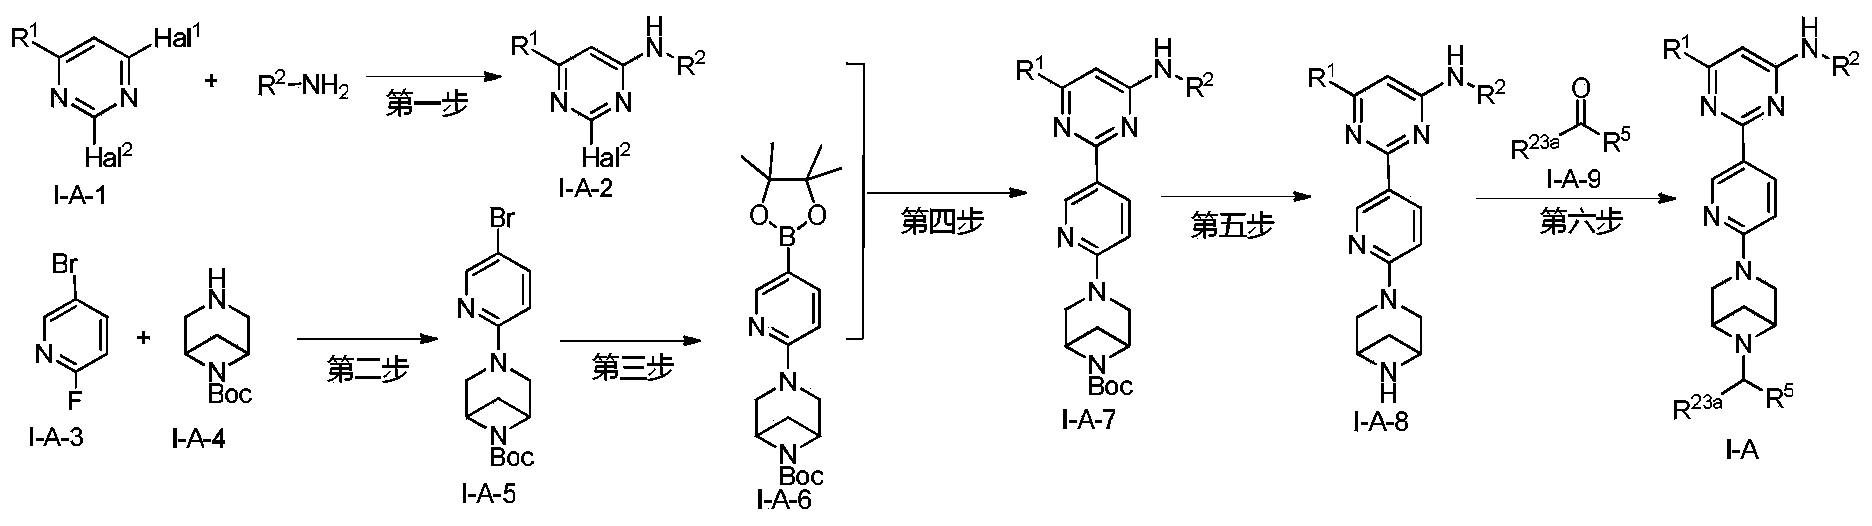
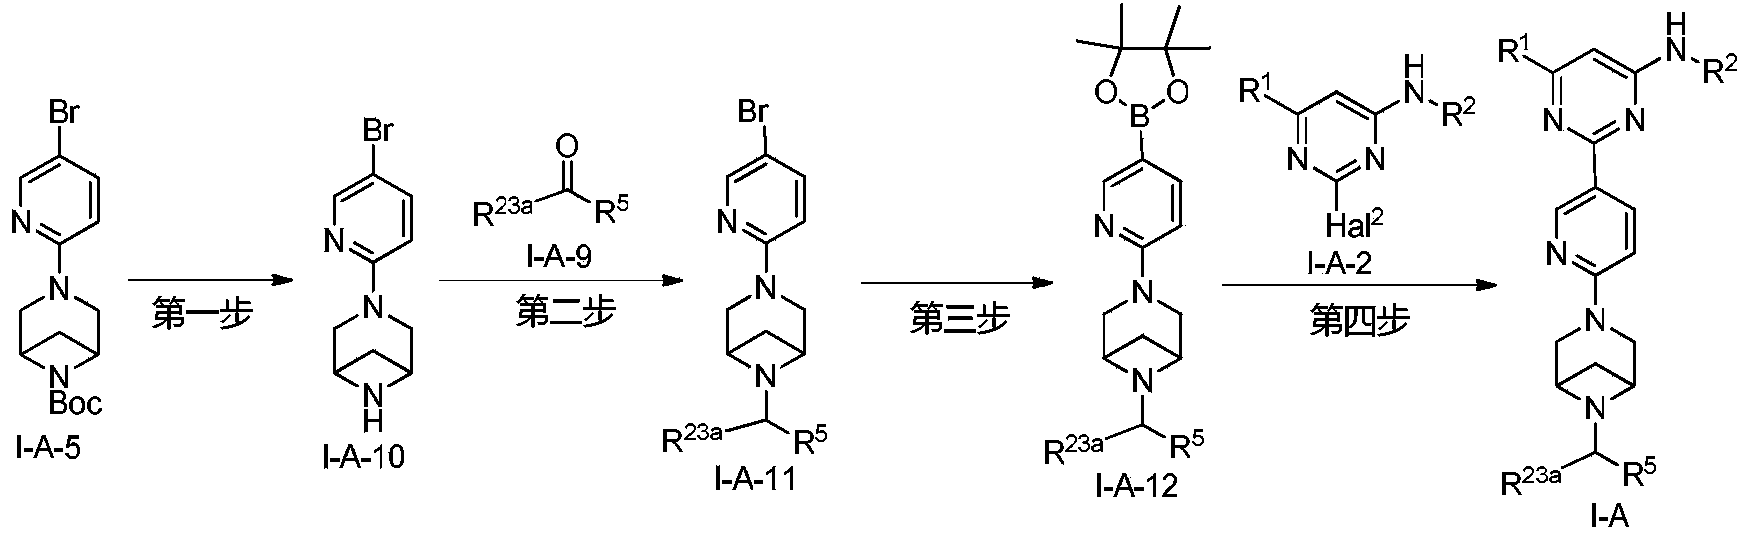
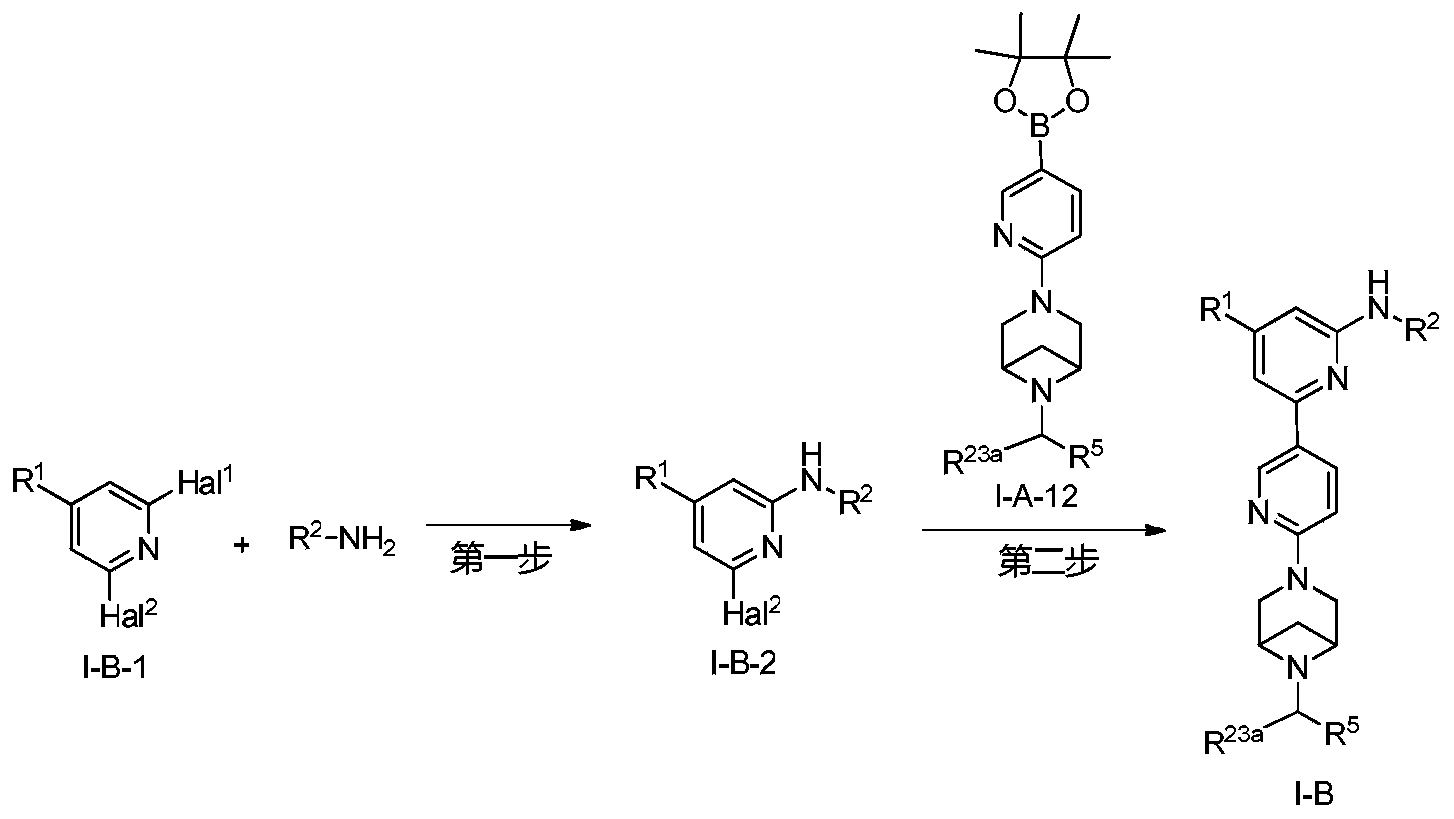
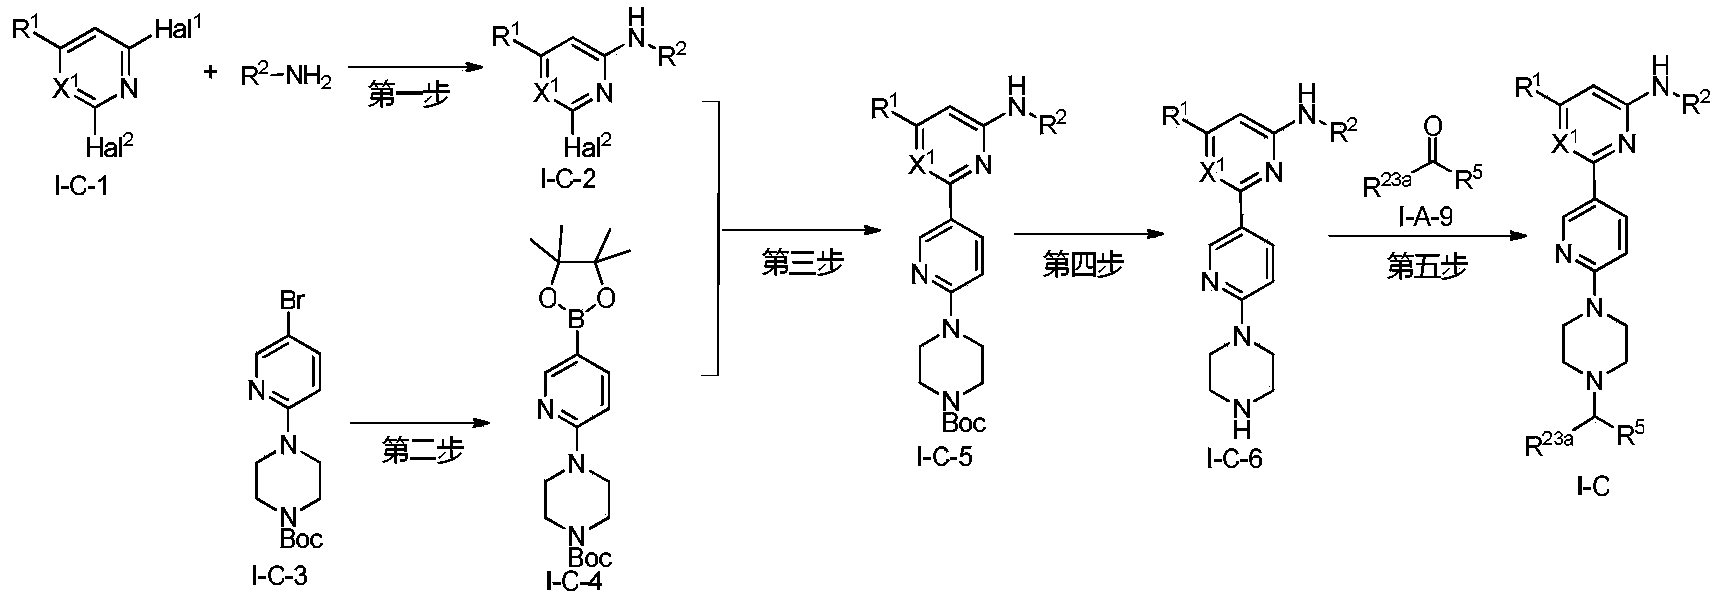
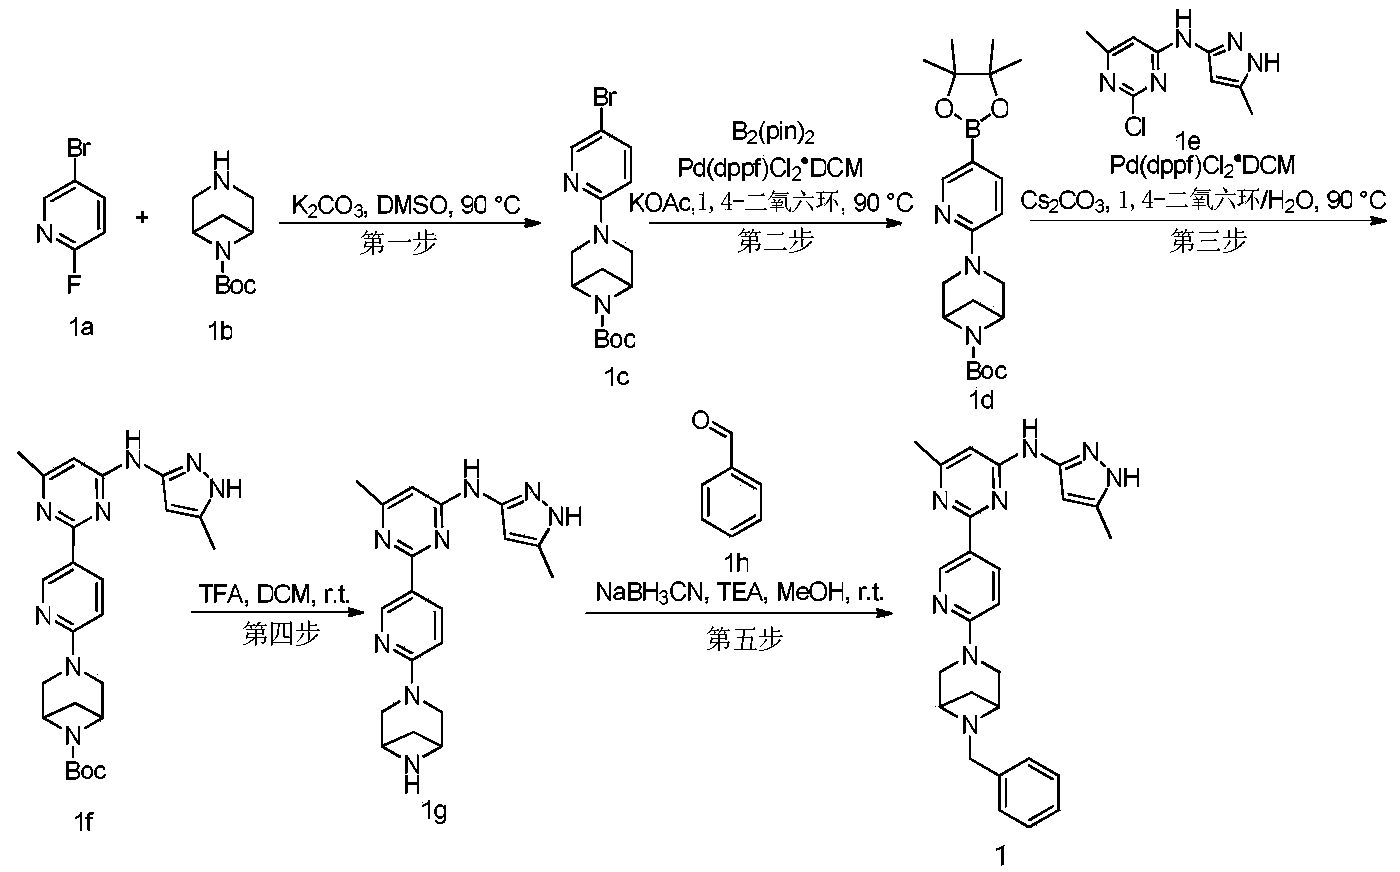
Example 6: 2-(6-(6-((6-(4-fluoro-1H-pyrazol-1-yl)pyridin-3-yl)methyl)-3,6-diazabicyclo[3.1.1]heptane-3-yl)pyridin-3-yl)-6-methyl-N-(5-methyl-1H-pyrazol-3-yl)pyrimidin-4-amine (Compound 17)
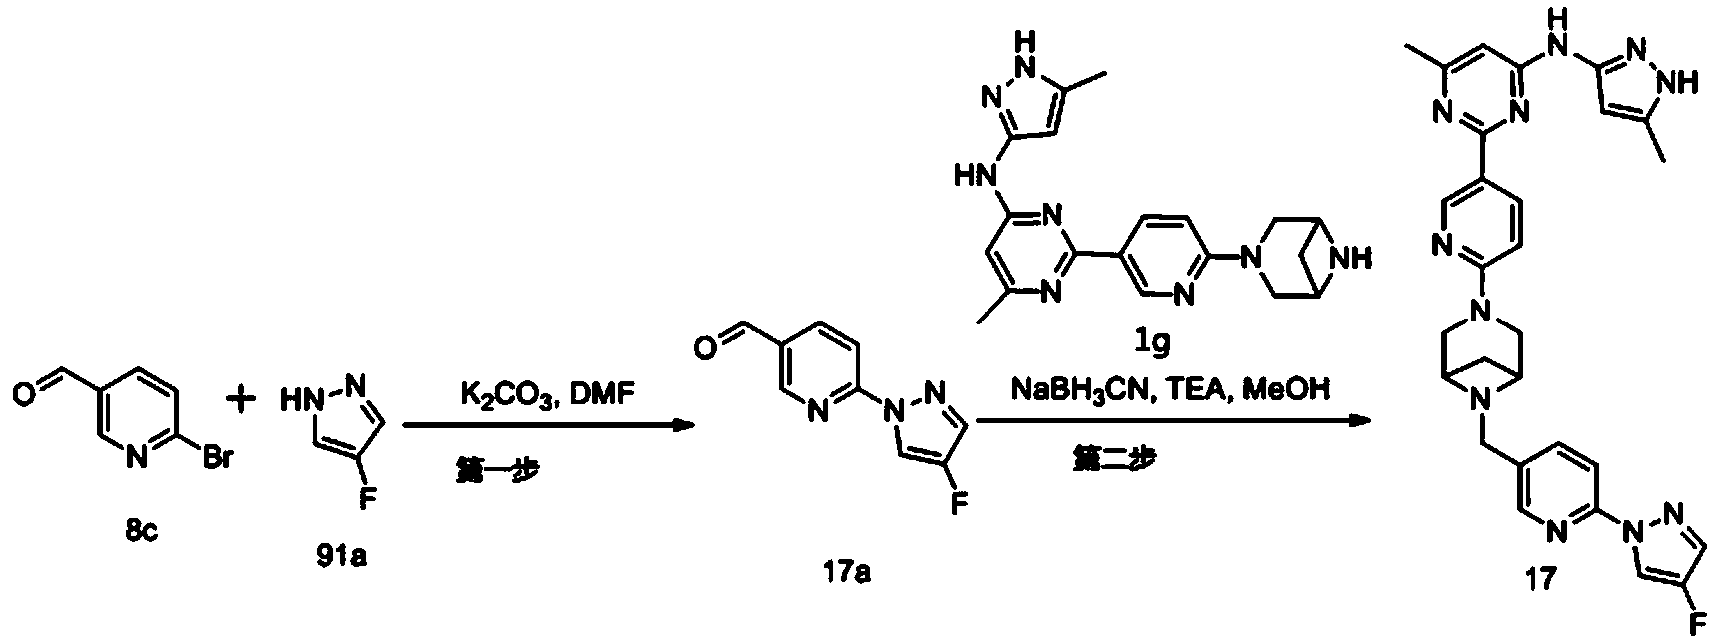
Step 1: Preparation of 6-(4-fluoro-1H-pyrazol-1-yl)nicotinaldehyde (compound 17a)
[0396]Compound 8c (2.0 g), 91a hydrochloride (1.58 g), and potassium carbonate (4.45 g) were sequentially added to DMF (15 mL), and the mixture was heated to 80 °C and stirred for 14 h. The reaction mixture was cooled to room temperature, diluted with water (100 mL), and extracted with DCM (50 mL x 2). The organic phases were combined, washed with water and saturated brine, dried over anhydrous sodium sulfate, filtered, concentrated under reduced pressure, and purified by silica gel column chromatography (PE:EA = 10:1) to give compound 17a (0.81 g). MS m/z (ESI): 192.1 [M+H]
[0397]Step 2: Preparation of 2-(6-(6-((6-(4-fluoro-1H-pyrazol-1-yl)pyridin-3-yl)methyl)-3,6-diazabicyclo[3.1.1]heptane-3-yl)pyridin-3-yl)-6-methyl-N-(5-methyl-1H-pyrazol-3-yl)pyrimidin-4-amine (compound 17)
[0398]1 g of trifluoroacetate (22.82 mg) and compound 17a (27.47 mg) were added to methanol (1.0 mL), followed by the sequential addition of triethylamine (4.45 mg) and sodium cyanoborohydride (13.86 mg), and the reaction was carried out at room temperature for 14 h. After the reaction was completed, the reaction solution was concentrated to dryness under reduced pressure and purified by Prep-HPLC to obtain compound 17 (7.0 mg). MS m/z (ESI): 538.3 [M+H]
[0399]
1H NMR(400MHz,DMSO-d 6)δ11.98(s,1H),9.66(s,1H),9.12(d,J=2.16Hz,1H),8.67(dd,J=4.54,0.64Hz,1H),8.43(dd,J=8.94,2.28Hz,1H),8.41(d,J=1.68,1H),7.98(dd,J=8.48Hz,2.12 1H),7.92(d,J=4.28,1H),7.87(d,J=8.4,1H),6.78(d,J=9.0Hz,2H),6.31(br,1H),3.78-3.71(m,4H),3.68-3.52(m,4H),2.59-2.52(m,1H),2.33(s,3H),2.25(s,3H),1.60(d,J=8.36Hz,1H).
PAT
- Heterocyclic compound, pharmaceutical composition comprising same, preparation method therefor, and use thereofPublication Number: US-2022144847-A1Priority Date: 2019-02-19
- Heterocyclic compounds, pharmaceutical compositions containing the same and preparation methods and uses thereofPublication Number: CN-113316578-BPriority Date: 2019-02-19Grant Date: 2023-10-31
- Heterocyclic compounds, pharmaceutical compositions containing the same and preparation methods and uses thereofPublication Number: CN-117263945-APriority Date: 2019-02-19
- Heterocyclic compounds, pharmaceutical compositions containing the same and preparation methods and uses thereofPublication Number: CN-117327078-APriority Date: 2019-02-19
- Heterocyclic compounds, pharmaceutical compositions containing same, methods for their preparation and usePublication Number: JP-7615056-B2Priority Date: 2019-02-19Grant Date: 2025-01-16
- Salt and crystal form of pyrimidine compound, and preparation methods thereforPublication Number: US-2023295174-A1Priority Date: 2020-07-28
- Heterocyclic compound, pharmaceutical composition comprising same, preparation method therefor, and use thereofPublication Number: WO-2020168939-A1Priority Date: 2019-02-19
- Heterocyclic compounds, pharmaceutical compositions containing the same, and preparation methods and uses thereofPublication Number: CN-113316578-APriority Date: 2019-02-19
- Heterocyclic compound, pharmaceutical composition comprising same, preparation method therefor, and use thereofPublication Number: EP-3929198-A1Priority Date: 2019-02-19
- Heterocyclic compounds, drug compositions containing them, methods of their manufacture and usePublication Number: JP-2022521859-APriority Date: 2019-02-19
- Use of heterocyclic compound for treating diseases related to ret genetic change and method thereforPublication Number: WO-2024240017-A1Priority Date: 2023-05-19
- Uses and methods of heterocyclic compounds for treating diseases associated with kinase resistance mutationsPublication Number: CN-116801882-APriority Date: 2021-03-24
- Use of heterocyclic compound in treating diseases related to kinase drug-resistant mutation and method thereforPublication Number: EP-4316490-A1Priority Date: 2021-03-24
- Salt and crystal form of pyrimidine compound, and preparation methods thereforPublication Number: EP-4190781-A1Priority Date: 2020-07-28
- Salts, crystal forms of pyrimidine compounds and methods for their preparationPublication Number: JP-2023535361-APriority Date: 2020-07-28
AS ON OCT2025 4.511 LAKHS VIEWS ON BLOG WORLDREACH AVAILABLEFOR YOUR ADVERTISEMENT
join me on Linkedin
Anthony Melvin Crasto Ph.D – India | LinkedIn
join me on Researchgate
RESEARCHGATE
join me on Facebook
Anthony Melvin Crasto Dr. | Facebook
join me on twitter
Anthony Melvin Crasto Dr. | twitter
+919321316780 call whatsaapp
EMAIL. amcrasto@gmail.com
……
//////////Lunbotinib, tyrosine kinase inhibitor, antineoplastic, KL3T9ZU6HQ
Lirodegimod
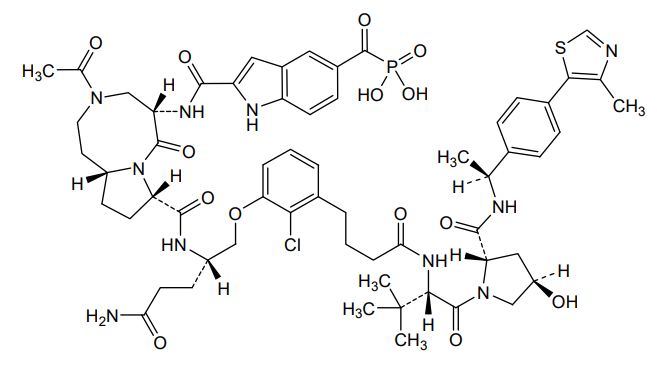
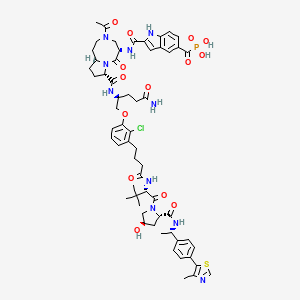
Lirodegimod
CAS 2502186-79-8
MF C60H74ClN10O14PS, MW 1257.79
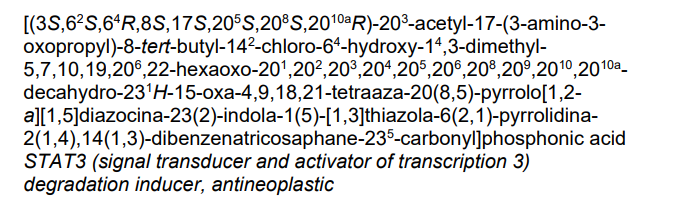
[2-[[(5S,8S,10aR)-3-acetyl-8-[[(2S)-5-amino-1-[2-chloro-3-[4-[[(2S)-1-[(2S,4R)-4-hydroxy-2-[[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]carbamoyl]pyrrolidin-1-yl]-3,3-dimethyl-1-oxobutan-2-yl]amino]-4-oxobutyl]phenoxy]-5-oxopentan-2-yl]carbamoyl]-6-oxo-1,2,4,5,8,9,10,10a-octahydropyrrolo[1,2-a][1,5]diazocin-5-yl]carbamoyl]-1H-indole-5-carbonyl]phosphonic acid
KT 333, KT333, ANTINEOPLASTIC, Fast Track (United States), Orphan Drug (United States), 4Q6ZHJ2MNA
Lirodegimod is a small molecule drug. The usage of the INN stem ‘-imod’ in the name indicates that Lirodegimod is a immunomodulator, both stimulant/suppressive and stimulant. Lirodegimod has a monoisotopic molecular weight of 1256.45 Da.
Safety, PK, PD, Clinical Activity of KT-333 in Adult Patients With Refractory Lymphoma, Large Granular Lymphocytic Leukemia, Solid Tumors
CTID: NCT05225584
Phase: Phase 1
Status: Completed
Date: 2025-03-19
PAT
- Stat3 degraders and uses thereofPublication Number: US-2023212201-A1Priority Date: 2021-12-11
- Stat3 degraders and uses thereofPublication Number: US-2025019388-A1Priority Date: 2021-12-11
- Stat degraders and uses thereofPublication Number: US-2024016942-A1Priority Date: 2020-03-17
- Stat degraders and uses thereofPublication Number: WO-2020206424-A1Priority Date: 2019-04-05
- Stat degraders and uses thereofPublication Number: US-11746120-B2Priority Date: 2019-04-05Grant Date: 2023-09-05
AS ON OCT2025 4.511 LAKHS VIEWS ON BLOG WORLDREACH AVAILABLEFOR YOUR ADVERTISEMENT
join me on Linkedin
Anthony Melvin Crasto Ph.D – India | LinkedIn
join me on Researchgate
RESEARCHGATE
join me on Facebook
Anthony Melvin Crasto Dr. | Facebook
join me on twitter
Anthony Melvin Crasto Dr. | twitter
+919321316780 call whatsaapp
EMAIL. amcrasto@gmail.com
……
///////////Lirodegimod, KT 333, KT333, ANTINEOPLASTIC, Fast Track, Orphan Drug, 4Q6ZHJ2MNA
Inlexisertib
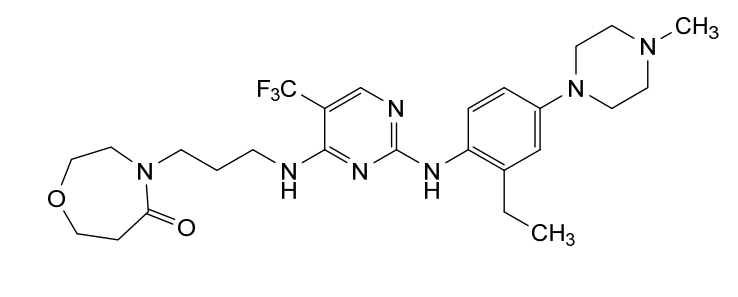
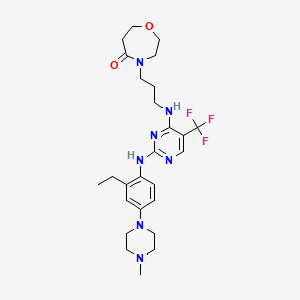
Inlexisertib
CAS 2543673-19-2
MF C26H36F3N7O2, 535.62
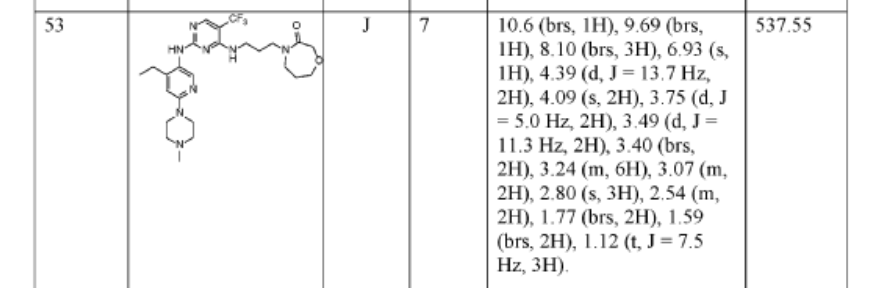
4-(3-((2-((2-ethyl-4-(4-methylpiperazin-1-yl)phenyl)amino)-5-(trifluoromethyl)pyrimidin-4-yl)amino)propyl)-1,4-oxazepan-5-one
4-[3-[[2-[2-ethyl-4-(4-methylpiperazin-1-yl)anilino]-5-(trifluoromethyl)pyrimidin-4-yl]amino]propyl]-1,4-oxazepan-5-one
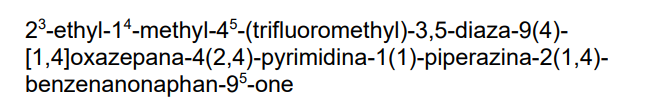
serine/ threonine kinase inhibitor, antineoplastic, DCC 3116, JM2ZTM8S7S
Inlexisertib is an orally bioavailable inhibitor of the serine/threonine-protein kinase ULK 1 and 2, with potential antineoplastic activity. Upon oral administration, inlexisertib targets and binds to ULK1/2. This inhibits cancer autophagy, which mutant RAS cancer cells use for their survival, and results in tumor cell death. ULK1/2 mediates the autophagocytotic process and is often upregulated in cancers, especially in mutant RAS cancers. Autophagy plays a key role in a tumor cell proliferation and survival, and mediates tumor cell resistance.
- A Study of Inlexisertib (DCC-3116) in Combination With Anticancer Therapies in Participants With Advanced MalignanciesCTID: NCT05957367Phase: Phase 1/Phase 2Status: RecruitingDate: 2025-06-05
- A Phase 1/2 Study of Inlexisertib (DCC-3116) in Patients With RAS/MAPK Pathway Mutant Solid TumorsCTID: NCT04892017Phase: Phase 1/Phase 2Status: RecruitingDate: 2025-05-06
SYN
https://patents.google.com/patent/US11530206B2/en
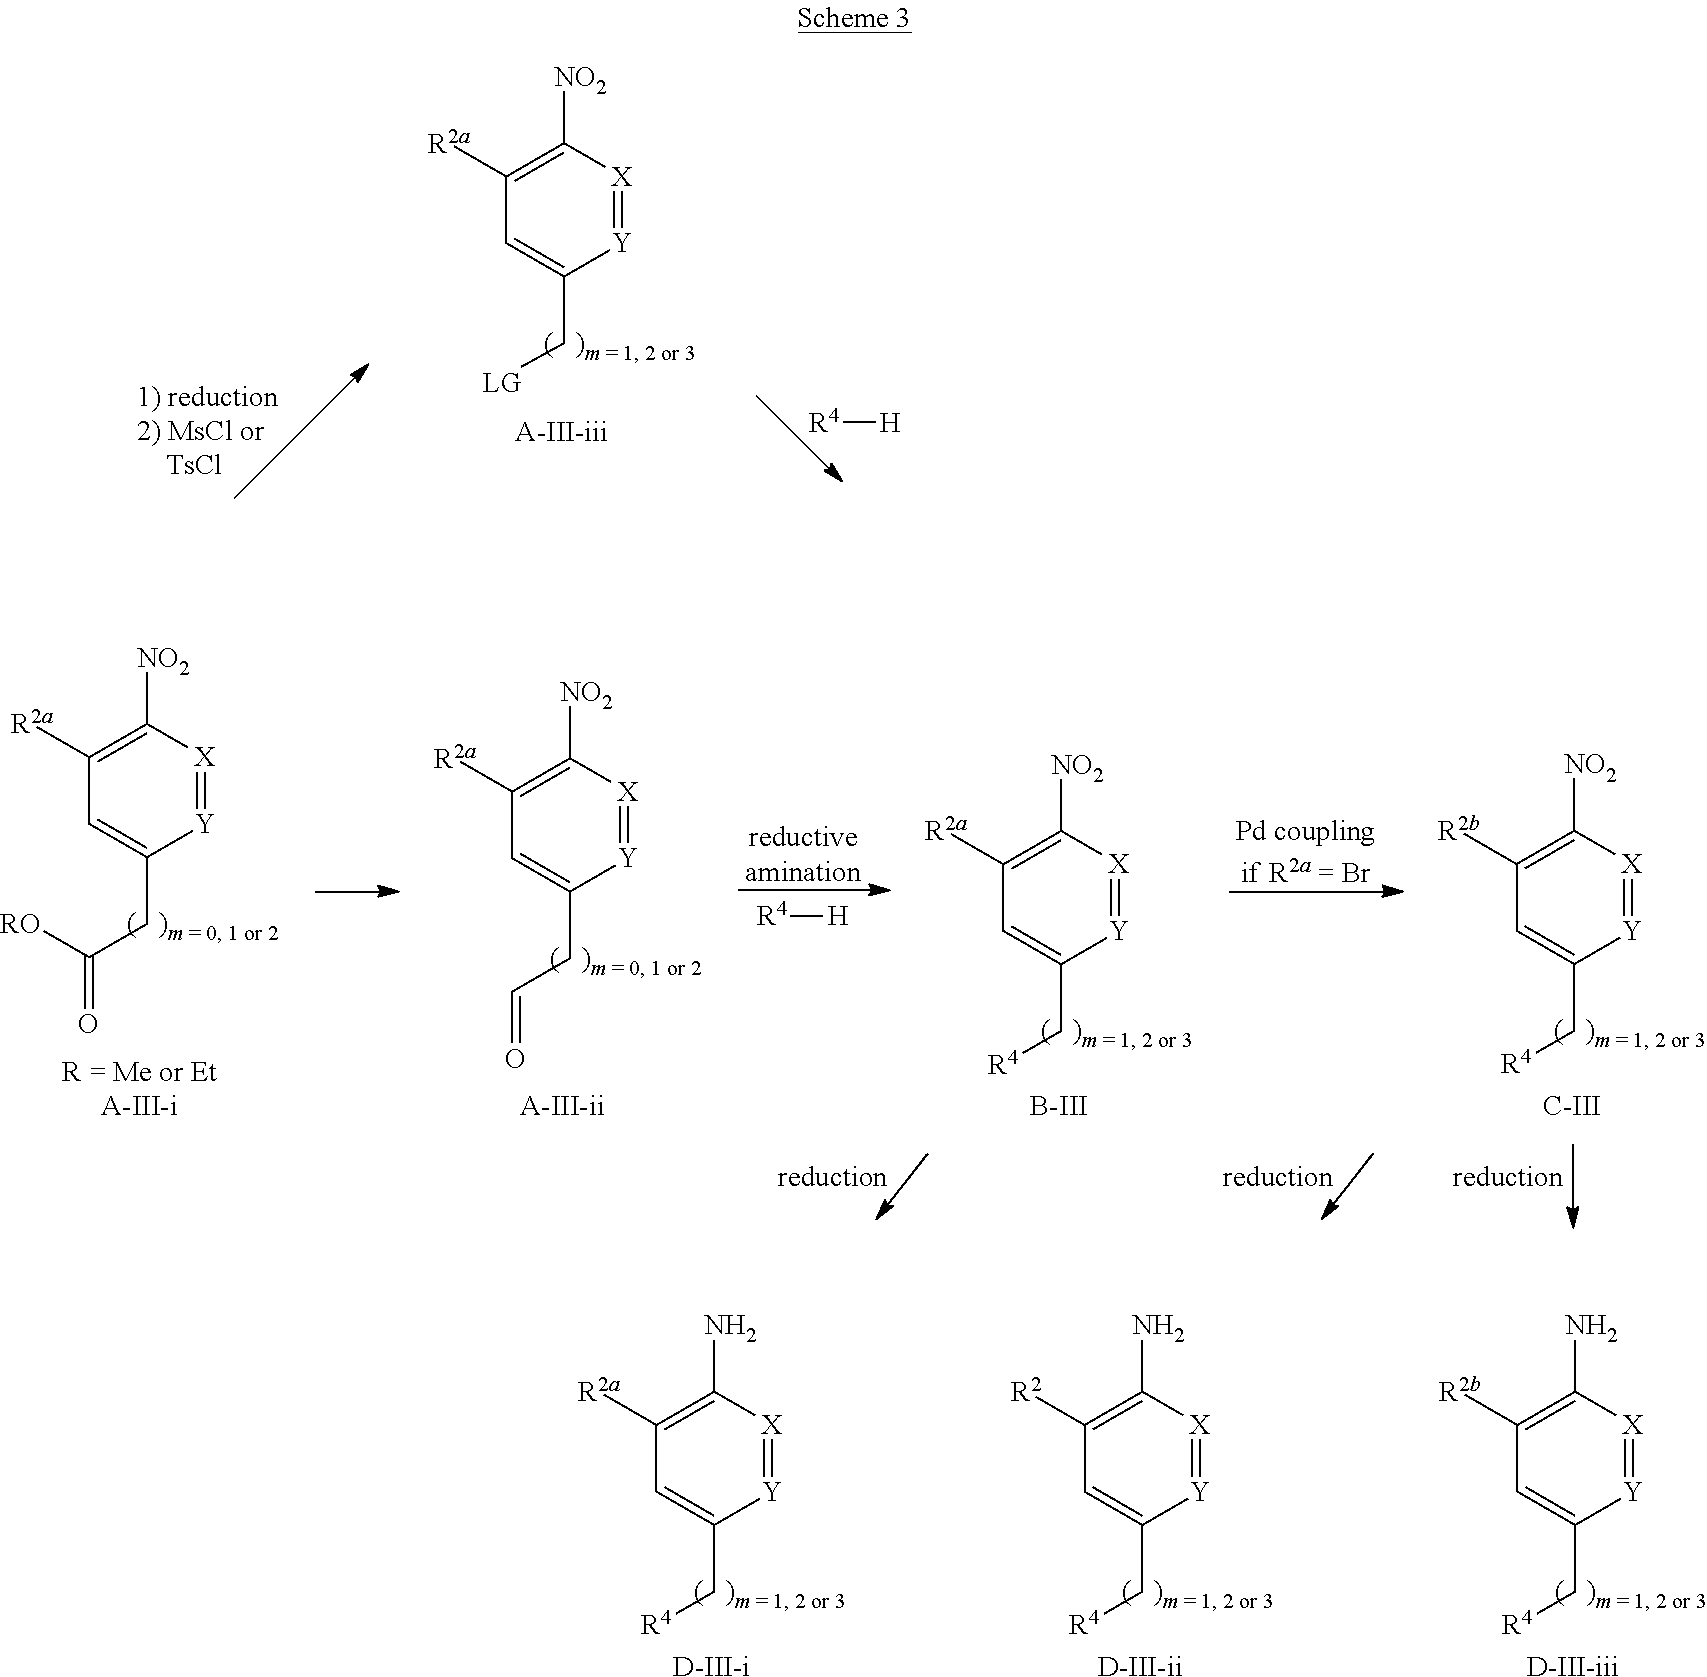
PAT
Phenylaminopyrimidine amide autophagy inhibitors and methods of use thereof
Publication Number: JP-7593947-B2
Priority Date: 2019-05-10
Grant Date: 2024-12-03
- PHENYLAMINOPYRIMIDINE AMIDE INHIBITORS OF AUTOPHAGY AND METHODS OF THEIR APPLICATIONPublication Number: HR-P20231730-T1Priority Date: 2019-05-10
- Phenylaminopyrimidine amide autophagy inhibitors and methods of use thereofPublication Number: US-12071432-B2Priority Date: 2019-05-10Grant Date: 2024-08-27
- Phenylaminopyrimidine amide autophagy inhibitors and methods of use thereofPublication Number: CN-118878519-APriority Date: 2019-05-10
- Phenylaminopyrimidine amide autophagy inhibitors and methods of use thereofPublication Number: CN-118878520-APriority Date: 2019-05-10
- Phenylaminopyrimidine amide autophagy inhibitors and methods of use thereofPublication Number: CN-118930524-APriority Date: 2019-05-10
- Phenylaminopyrimidine amide autophagy inhibitors and methods of use thereofPublication Number: AU-2020275392-B2Priority Date: 2019-05-10Grant Date: 2023-09-14
- Phenylaminopyrimidine amide autophagy inhibitors and methods of use thereofPublication Number: CN-114127057-BPriority Date: 2019-05-10Grant Date: 2024-07-12
- Phenylaminopyrimidine amide autophagy inhibitors and methods of use thereofPublication Number: EP-3966207-B1Priority Date: 2019-05-10Grant Date: 2023-11-01
- Phenylaminopyrimidine amide autophagy inhibitors and methods of use thereofPublication Number: EP-4342469-A2Priority Date: 2019-05-10
- Phenylaminopyrimidine amide autophagy inhibitors and methods of use thereofPublication Number: ES-2966807-T3Priority Date: 2019-05-10Grant Date: 2024-04-24
- Phenylaminopyrimidine amide autophagy inhibitors and methods of use thereofPublication Number: EP-3966207-A1Priority Date: 2019-05-10
- Phenylaminopyrimidine amide autophagy inhibitors and methods of use thereofPublication Number: KR-20220008873-APriority Date: 2019-05-10
- Phenylaminopyrimidine amide autophagy inhibitor and how to use itPublication Number: JP-2022531801-APriority Date: 2019-05-10
- Phenylaminopyrimidine amide autophagy inhibitors and methods of use thereofPublication Number: US-11530206-B2Priority Date: 2019-05-10Grant Date: 2022-12-20
- Phenylaminopyrimidine amide autophagy inhibitors and methods of use thereofPublication Number: US-2023039712-A1Priority Date: 2019-05-10
- Combination of dcc-3116 and mapkap pathway inhibitors for use in the treatment of cancerPublication Number: WO-2024050351-A1Priority Date: 2022-09-02
- Phenylaminopyrimidine amide autophagy inhibitors and methods of use thereofPublication Number: US-2020354352-A1Priority Date: 2019-05-10
- Phenylaminopyrimidine amide autophagy inhibitors and methods of use thereofPublication Number: WO-2020231806-A1Priority Date: 2019-05-10
- Phenylaminopyrimidine amide autophagy inhibitors and methods of use thereofPublication Number: AU-2020275392-A1Priority Date: 2019-05-10
- Phenylaminopyrimidine amide autophagy inhibitor and method of usePublication Number: CN-114127057-APriority Date: 2019-05-10
PAT
PAT
https://patentscope.wipo.int/search/en/detail.jsf?docId=WO2020231806&_cid=P12-MHCSWS-98394-1

AS ON JUNE2025 4.45 LAKHS VIEWS ON BLOG WORLDREACH AVAILABLEFOR YOUR ADVERTISEMENT
join me on Linkedin
Anthony Melvin Crasto Ph.D – India | LinkedIn
join me on Researchgate
RESEARCHGATE
join me on Facebook
Anthony Melvin Crasto Dr. | Facebook
join me on twitter
Anthony Melvin Crasto Dr. | twitter
+919321316780 call whatsaapp
EMAIL. amcrasto@gmail.com
……
/////////Inlexisertib, serine/ threonine kinase inhibitor, antineoplastic, DCC 3116, JM2ZTM8S7S
Icovamenib
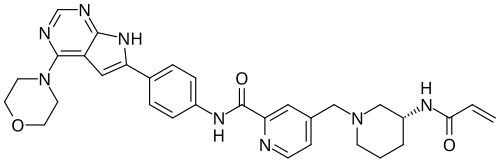
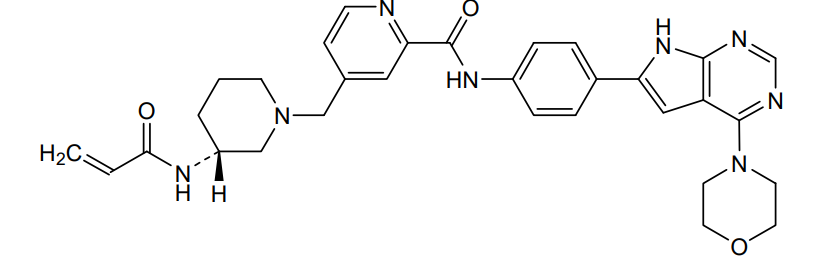
Icovamenib
CAS 2448172-22-1
MF C31H34N8O3 MW 566.7 g/mol
N-{4-[4-(morpholin-4-yl)-7H-pyrrolo[2,3-d]pyrimidin-6-yl]phenyl}-4-{[(3R)-3-(prop-2-enamido) piperidin-1-yl]methyl}pyridine-2-carboxamide
N-[4-(4-morpholin-4-yl-7H-pyrrolo[2,3-d]pyrimidin-6-yl)phenyl]-4-[[(3R)-3-(prop-2-enoylamino)piperidin-1-yl]methyl]pyridine-2-carboxamide
menin-MLL (mixed-lineage leukemia) protein interaction inhibitor,
antineoplastic, BMF-219, BMF 219, 2Z737MY35A, Menin-MLL inhibitor 21
Icovamenib is an investigational irreversible covalent inhibitor of menin. It is developed by Biomea Fusion for diabetes, lymphoma, leukemia, and multiple myeloma.[1][2][3]
Icovamenib is an orally bioavailable, irriversible inhibitor of menin, an essential co-factor of oncogenic menin-mixed lineage leukemia (MLL; myeloid/lymphoid leukemia; KMT2A) fusion proteins, with potential antineoplastic activity. Upon oral administration, icovamenib specifically targets and binds to menin, thereby preventing the interaction between the two proteins menin and MLL and the formation of the menin-MLL complex. This reduces the expression of downstream target genes, such as MYC and Bcl2, and results in an inhibition of the proliferation of MLL-rearranged tumor cells. Menin, an essential transcriptional regulator, plays a key role in oncogenic signaling in cancers driven by oncogenic MLL-fusions.
SYN
https://patentscope.wipo.int/search/en/detail.jsf?docId=US299042443&_cid=P20-MH9YDY-31032-1
Example 9
Synthesis of Compound 10
Compound 10
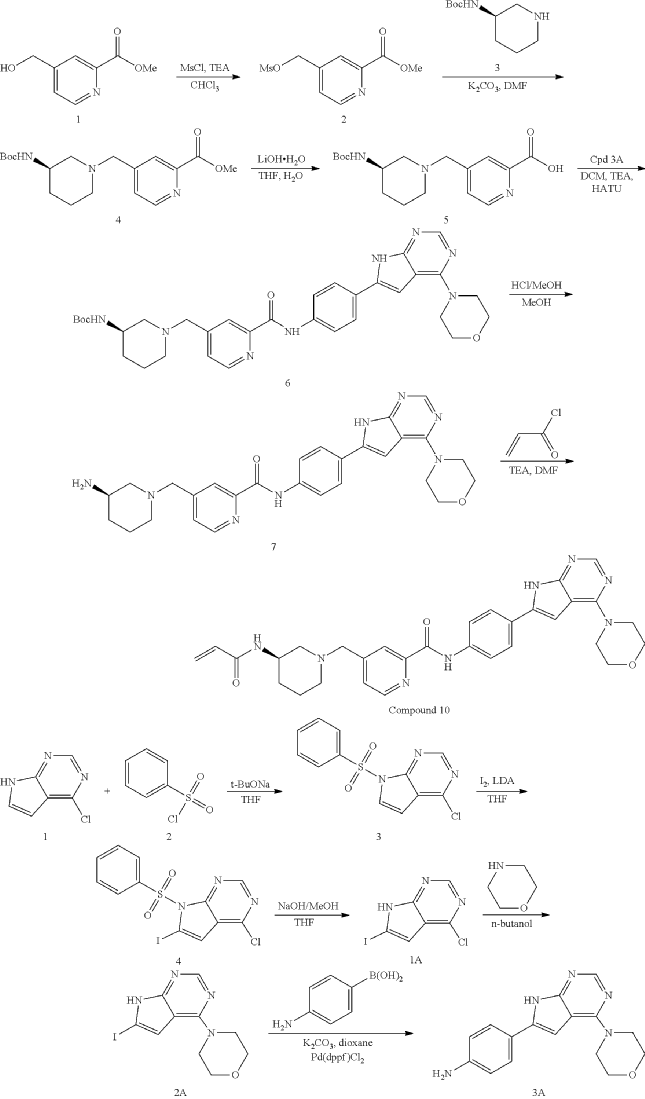
General Procedure for Preparation of Intermediate 2
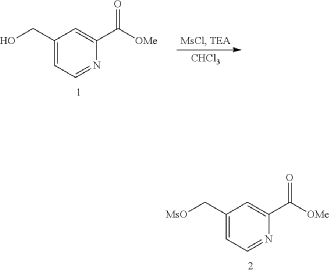
| 1H NMR: CDCl 3 400 MHz 8.80 (d, J=4.85 Hz, 1H), 8.15 (d, J=0.66 Hz, 1H), 7.53 (dt, J=4.91, 0.85 Hz, 1H), 5.27-5.34 (m, 2H), 4.00-4.08 (m, 3H), 3.11 (s, 3H) |
General Procedure for Preparation of Intermediate 5—
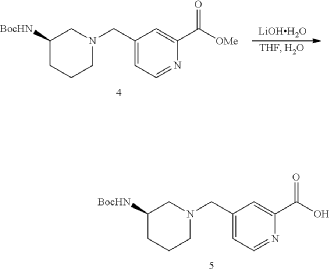
| To a solution of Intermediate 4 (1.50 g, 4.29 mmol, 1 eq) in THF (7.00 mL) was added LiOH.H 2O (540.3 mg, 12.8 mmol, 3 eq) in H 2O (7.00 mL). The mixture was stirred at 25° C. for 3 h. TLC (Dichloromethane:Methanol=10:1, R f=0) showed the reaction was complete. The mixture was poured into H 2O (20.0 mL) and extracted with DCM (10.0 mL×3). Then the organic phases dried over Na 2SO 4, filtered and concentrated under vacuum. The crude without purification. Give the Intermediate 5 (1.20 g, crude) as a yellow solid. |
General Procedure for Preparation of Intermediate 6—
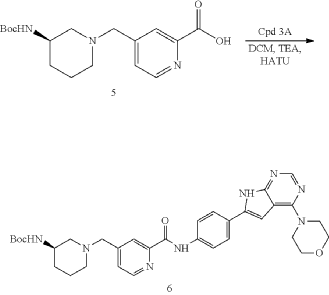
| To a solution of Intermediate 5 (0.80 g, 2.39 mmol, 1 eq), Intermediate 3A (704.4 mg, 2.39 mmol, 1 eq), TEA (1.69 g, 16.7 mmol, 2.32 mL, 7 eq) in DCM (10.0 mL) was added HATU (1.36 g, 3.58 mmol, 1.5 eq). The mixture was stirred at 20° C. for 12 h. LCMS showed the reaction was complete. The mixture was poured into H 2O (40.0 mL) and extracted with DCM (20.0 mL×3). Then the organic phases were washed with brine (50.0 mL) dried over Na 2SO 4, filtered and concentrated under vacuum. The crude for next step without purification. Give the Intermediate 6 (0.60 g, crude) as a yellow solid. |
General Procedure for Preparation of Intermediate 7—
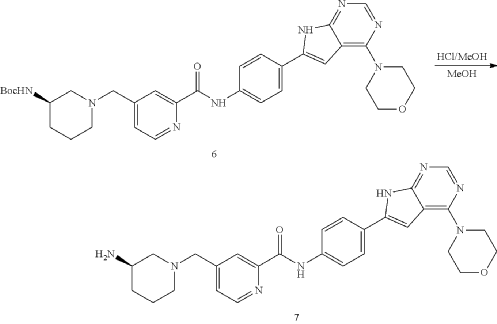
| To a solution of Intermediate 6 (0.50 g, 816.0 umol, 1 eq) in MeOH (5.00 mL) was added HCl/MeOH (4 M, 5.00 mL, 24.51 eq). The mixture was stirred at 20° C. for 12 h. LCMS showed the reaction was complete. The mixture was concentrated under vacuum. The crude for next step without purification. Give the Intermediate 7 (0.50 g, crude, HCl) as a yellow solid. |
General Procedure for Preparation of Compound 10—
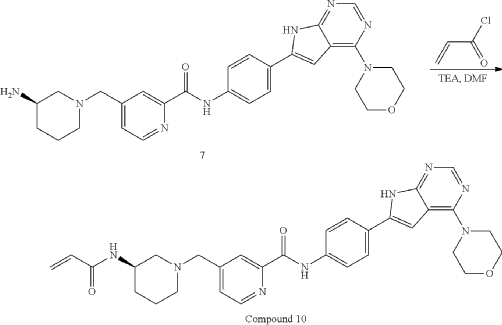
| To a solution of Intermediate 3 (0.50 g, 910.6 umol, 1 eq, HCl) in DMF (10.0 mL) was added TEA (645.0 mg, 6.37 mmol, 887.2 uL, 7 eq) and prop-2-enoyl chloride (82.4 mg, 910.6 umol, 74.2 uL, 1 eq). Then the mixture was stirred at 20° C. for 12 h. LCMS showed the reaction was complete. The mixture was poured into H 2O (50.0 mL), then was filtered and filter cake was concentrated in vacuum. The crude product was purified by reversed-phase HPLC (column: Phenomenex Luna C18 200*40 mm*10 um; mobile phase: [water(0.05% HCl)-ACN]; B %: 10%-30%, 10 min) and (column: Xtimate C18 150*25 mm*5 um; mobile phase: [water(10 mM NH 4HCO 3)-ACN]; B %: 30%-60%, 10 min). Give the Intermediate Compound 10 (20.0 mg, 35.0 umol, 3.85% yield, 99.3% purity) as a yellow solid. |
PAT
https://patentscope.wipo.int/search/en/detail.jsf?docId=WO2024172911&_cid=P20-MH9YNT-37455-1
PAT
- Substituted pyridines as irreversible inhibitors of menin-MLL interactionPublication Number: US-11702421-B2Priority Date: 2018-12-31Grant Date: 2023-07-18
- Irreversible inhibitors of menin-mll interactionPublication Number: US-2023227458-A1Priority Date: 2018-12-31
- N-[4-[4-(4-MORPHOLINYL)-7H-PYRROLO[2,3-d]PYRIMIDIN-6-YL]PHENYL]-4-[[3(R)-[(1-OXO-2-PROPEN-1-YL)AMINO]-1-PIPERIDINYL]METHYL]-2-PYRIDINE CARBOXAMIDE AND USES THEREOFPublication Number: US-2024376112-A1Priority Date: 2018-12-31
- Covalent inhibitors of menin-mll interaction for diabetes mellitusPublication Number: WO-2023018825-A1Priority Date: 2021-08-11
- Covalent inhibitors of Mennin-MLL interaction for diabetesPublication Number: CN-118076357-APriority Date: 2021-08-11
- Irreversible inhibitors of menin-mll interactionPublication Number: US-2020223853-A1Priority Date: 2018-12-31
- Substituted pyridines as irreversible inhibitors of menin-MLL interactionPublication Number: US-11084825-B2Priority Date: 2018-12-31Grant Date: 2021-08-10
- Irreversible inhibitors of menin-mll interactionPublication Number: US-2022169627-A1Priority Date: 2018-12-31
- Crystalline forms of an irreversible inhibitor of menin-mll interactionPublication Number: US-2023086137-A1Priority Date: 2021-08-20
- Crystalline form of n-[4-[4-(4-morpholinyl)-7h-pyrrolo[2,3-d]pyrimidin-6-yl]phenyl]-4-[[3(r)-[(1-oxo -2-propen-1-yl)amino]-1-piperidinyl]methyl]-2-pyridinecarboxamide, an irreversible menin-mll inhibitor for the treatment of cancerPublication Number: EP-4387972-A1Priority Date: 2021-08-20
- Crystalline forms of N-[4-[4-(4-morpholinyl)-7H-pyrrolo[2,3-d]pyrimidin-6-yl]phenyl]-4-[[3(r)-[(1-oxo-2-propen-1-yl)amino]-1-piperidinyl]methyl]-2-pyridinecarboxamide as an irreversible inhibitor of menin-MLL interactionPublication Number: US-12018032-B2Priority Date: 2021-08-20Grant Date: 2024-06-25
- Crystalline form of n-[4-[4-(4-morpholinyl)-7h-pyrrolo[2,3-d]pyrimidin-6-yl]phenyl]-4-[[3(r)-[(1-oxo -2-propen-1-yl)amino]-1-piperidinyl]methyl]-2-pyridinecarboxamide, an irreversible menin-mll inhibitor for the treatment of cancerPublication Number: WO-2023022912-A1Priority Date: 2021-08-20
- Crystalline forms of an irreversible inhibitor of menin-mll interactionPublication Number: US-2024343731-A1Priority Date: 2021-08-20
- Synthetic methods for preparing a pyridinecarboxamide compoundPublication Number: WO-2024011450-A1Priority Date: 2022-07-13
- Menin-mll inhibitors and compositions for proliferation of beta cellsPublication Number: WO-2024006391-A1Priority Date: 2022-06-28
- Flt3 combination therapy for cancer and compositions thereforPublication Number: WO-2023225005-A1Priority Date: 2022-05-17
- Treatment of cancer with menin inhibitors and immuno-oncology agentsPublication Number: WO-2023172925-A1Priority Date: 2022-03-08
- Treatment of hematological malignancies with menin inhibitors and p-glycoprotein inhibitorsPublication Number: WO-2023150635-A1Priority Date: 2022-02-04
- Crystalline forms of N[4[4-(4-Morpholinyl)-7H-Pyrrolo[2-3-D]Pyrimidin-6-yl]Phenyl]-4-[[3(R)-[(1-Oxo-2-Protein-1-yl)Amino]-1-Piperidinyl]Methyl]2-Pyridinecarboxamide]Publication Number: US-12215113-B2Priority Date: 2023-01-18Grant Date: 2025-02-04
- CRYSTALLINE FORMS OF N-[4-[4-(4-MORPHOLINYL)-7H-PYRROLO[2,3-d]PYRIMIDIN-6-YL]PHENYL]-4-[[3(R)-[(1-OXO-2-PROPEN-1-YL)AMINO]-1-PIPERIDINYL]METHYL]-2-PYRIDINECARBOXAMIDE AS IRREVERSIBLE INHIBITORS OF MENIN-MLL INTERACTIONPublication Number: US-2024417404-A1Priority Date: 2023-01-18
- Crystalline forms of n-[4-[4-(4-morpholinyl)-7h-pyrrolo[2,3-d]pyrimidin-6- yl]phenyl]-4-[[3(r)-[(l-oxo-2-propen-l-yl)amino]-l-piperidinyl]methyl]-2-pyridinecarboxamide as a covalent inhibitor of menin-mll interactionPublication Number: WO-2024155710-A1Priority Date: 2023-01-18
- Crystalline forms of n-[4-[4-(4-morpholinyl)-7h-pyrrolo[2,3-d]pyrimidin-6-yl]phenyl]-4-[[3(r)-[(l-oxo-2-propen-l-yl)amino]-l-piperidinyl]methyl]-2- pyridinecarboxamide as a covalentinhibitor of menin-mll interactionPublication Number: WO-2024155719-A1Priority Date: 2023-01-18
- Combinations of lsd1 inhibitors and menin inhibitors for treating cancerPublication Number: WO-2024110649-A1Priority Date: 2022-11-24
AS ON JUNE2025 4.45 LAKHS VIEWS ON BLOG WORLDREACH AVAILABLEFOR YOUR ADVERTISEMENT
join me on Linkedin
Anthony Melvin Crasto Ph.D – India | LinkedIn
join me on Researchgate
RESEARCHGATE
join me on Facebook
Anthony Melvin Crasto Dr. | Facebook
join me on twitter
Anthony Melvin Crasto Dr. | twitter
+919321316780 call whatsaapp
EMAIL. amcrasto@gmail.com
……
References
- Rodriguez, Jose E.; Abitbol, Alexander; Abuzgaya, Fathi; Perez, Cesar; Mourya, Sanchita; Munneke, Brian; Morris, Stephan W.; Butler, Thomas (20 June 2023). “91-LB: COVALENT-111, a Phase 1/2 Trial of BMF-219, a Covalent Menin Inhibitor, in Patients with Type 2 Diabetes Mellitus—Preliminary Results”. Diabetes. 72 (Supplement_1) 91-LB. doi:10.2337/db23-91-LB. S2CID 259444592.
- Ravandi-Kashani, F.; Kishtagari, A.; Carraway, H.; Schiller, G.; Curran, E.; Yadav, B.; Cacovean, A.; Morris, S.; Butler, T.; Lancet, J. (23 June 2022). “P587: Covalent-101: A Phase 1 Study of BMF-219, A Novel Oral Irreversible Menin Inhibitor, in Patients with Relapsed/Refractory Acute Leukemia, Diffuse Large B-Cell Lymphoma, and Multiple Myeloma”. HemaSphere. 6: 486–487. doi:10.1097/01.HS9.0000845236.32931.83.
- Somanath, Priyanka; Lu, Daniel; Law, Brian; Archer, Tenley C.; Cacovean, Alexandru; Palmer, James T.; Kinoshita, Taisei; Butler, Thomas (5 November 2021). “Novel Irreversible Menin Inhibitor, BMF-219, Shows Potent Single Agent Activity in Clinically Relevant DLBCL Cells”. Blood. 138 (Supplement 1): 4318. doi:10.1182/blood-2021-148045.
| Clinical data | |
|---|---|
| Other names | BMF-219 |
| Legal status | |
| Legal status | Investigational |
| Identifiers | |
| IUPAC name | |
| CAS Number | 2448172-22-1 |
| PubChem CID | 154988914 |
| ChemSpider | 115037287 |
| UNII | 2Z737MY35A |
| Chemical and physical data | |
| Formula | C31H34N8O3 |
| Molar mass | 566.666 g·mol−1 |
| 3D model (JSmol) | Interactive image |
| SMILES | |
| InChI | |
/////////Icovamenib, antineoplastic, BMF-219, BMF 219, 2Z737MY35A, Menin-MLL inhibitor 21
Foselutoclax
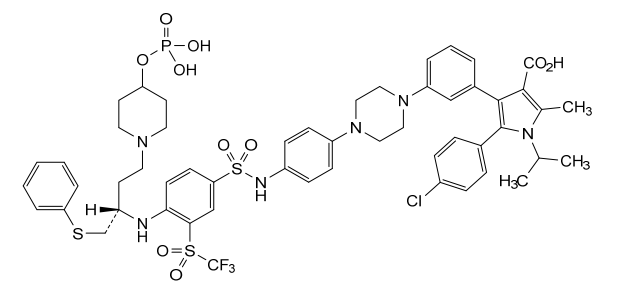
Foselutoclax
CAS 2271269-01-1
MF C53H59ClF3N6O10PS3 MW 1159.7 g/mol
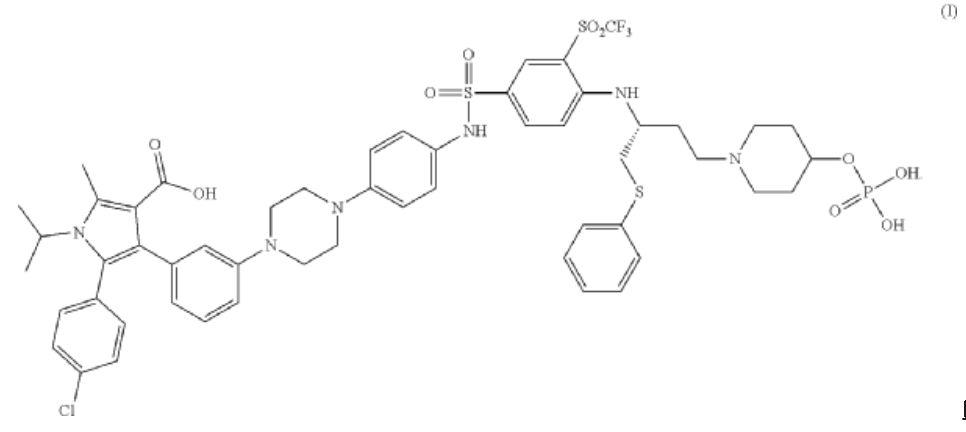
(10R)-14-chloro-25-methyl-7,7-dioxo-10-[(phenylsulfanyl)methyl]-134-(phosphonooxy)-21-(propan-2-yl)-83-(trifluoromethanesulfonyl)-21H-7λ6-thia-6,9-diaza-4(1,4)-piperazina-13(1)-piperidina-2(2,3)-pyrrola-1(1),3(1,3),5,8(1,4)-tetrabenzenatridecaphane-24-carboxylic acid
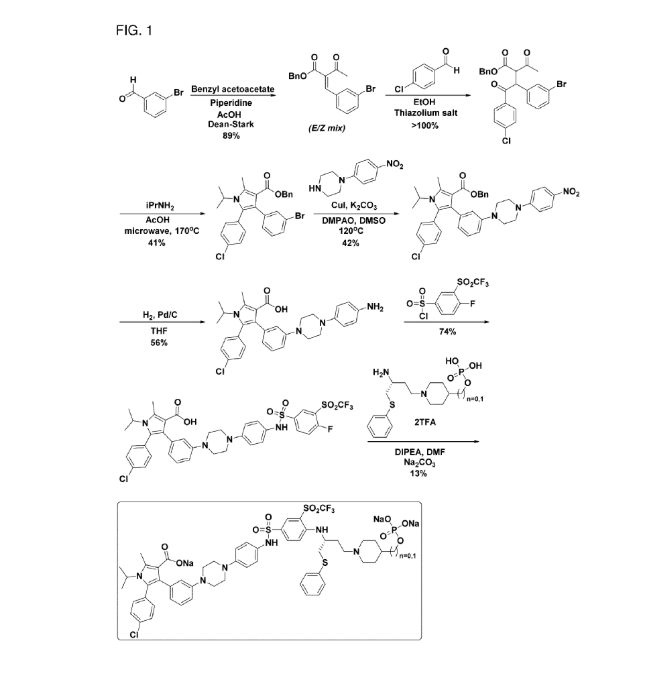
5-(4-chlorophenyl)-2-methyl-4-[3-[4-[4-[[4-[[(2R)-1-phenylsulfanyl-4-(4-phosphonooxypiperidin-1-yl)butan-2-yl]amino]-3-(trifluoromethylsulfonyl)phenyl]sulfonylamino]phenyl]piperazin-1-yl]phenyl]-1-propan-2-ylpyrrole-3-carboxylic acid
B-cell lymphoma 2 (Bcl-2) inhibitor, antineoplastic, VT53CL5GES, UBX 1325
Foselutoclax is an investigational new drug that is being evaluated for the treatment of age-related eye diseases, particularly diabetic macular edema (DME) and wet age-related macular degeneration (AMD). Developed by Unity Biotechnology, this senolytic compound acts as a potent inhibitor of Bcl-xL, a protein that senescent cells rely on for survival.[1] Foselutoclax is designed to selectively eliminate senescent cells in the retina, potentially addressing the underlying causes of vision loss in these conditions.[2]
- Assess the Efficacy and Safety of Repeat Intravitreal Injections of Foselutoclax (UBX1325) in Patients With DME (ASPIRE)CTID: NCT06011798Phase: Phase 2Status: CompletedDate: 2025-08-05
- Safety, Tolerability and Evidence of Activity Study of UBX1325 in Patients With Diabetic Macular Edema (BEHOLD)CTID: NCT04857996Phase: Phase 2Status: CompletedDate: 2024-05-16
- Safety and Tolerability Study of UBX1325 in Patients With Diabetic Macular Edema or Neovascular Age-Related Macular DegenerationCTID: NCT04537884Phase: Phase 1Status: CompletedDate: 2022-03-10
REF
- Therapeutic targeting of cellular senescence in diabetic macular edema: preclinical and phase 1 trial resultsPublication Name: Nature MedicinePublication Date: 2024-02PMID: 38321220DOI: 10.1038/s41591-024-02802-4
- Senolytics in the treatment of diabetic retinopathyPublication Name: Frontiers in PharmacologyPublication Date: 2022-08-26PMCID: PMC9462063PMID: 36091769DOI: 10.3389/fphar.2022.896907
- Senolytic drugs: from discovery to translationPublication Name: Journal of Internal MedicinePublication Date: 2020-08-04PMCID: PMC7405395PMID: 32686219DOI: 10.1111/joim.13141
- bcl-x, a bcl-2-related gene that functions as a dominant regulator of apoptotic cell deathPublication Name: CellPublication Date: 1993-08-27PMID: 8358789DOI: 10.1016/0092-8674(93)90508-n
PAT
Treatment of Lung Diseases Using Pharmaceutical Agents that Eliminate Senescent Cells
Publication Number: US-2020354336-A9
Priority Date: 2017-08-11
- Senescent Cells and for Treating CancerPublication Number: US-2022017485-A1Priority Date: 2018-06-13
- Acyl sulfonamides that are bcl family antagonists for use in clinical management of conditions caused or mediated by senescent cells and for treating cancerPublication Number: EP-4335516-A2Priority Date: 2018-06-13
- Methods of Inhibiting Pathological AngiogenesisPublication Number: US-2020253991-A1Priority Date: 2017-10-31
- Methods of inhibiting pathological angiogenesisPublication Number: US-11129838-B2Priority Date: 2017-10-31Grant Date: 2021-09-28
- Treatment of Lung Diseases Using Pharmaceutical Agents that Eliminate Senescent CellsPublication Number: US-2020199103-A1Priority Date: 2017-08-11
SYN
https://patentscope.wipo.int/search/en/detail.jsf?docId=US279621490&_cid=P21-MGPXU3-15237-1
SYN
https://patentscope.wipo.int/search/en/detail.jsf?docId=US421382898&_cid=P21-MGPXWE-19244-1
A crystalline solid meglumine salt of of (R)-5-(4-chlorophenyl)-1-isopropyl-2-methyl-4-(3-(4-(4-((4-((1-(phenylthio)-4-(4-((phosphonooxy)methyl)piperidin-1-yl)butan-2-yl)amino)-3-((trifluoromethyl)sulfonyl)phenyl)sulfonamido)phenyl)piperazin-1-yl)phenyl)-1H-pyrrole-3-carboxylic acid, the compound of Formula I:
SYN
https://patentscope.wipo.int/search/en/detail.jsf?docId=US348024244&_cid=P21-MGPXWE-19244-1
AS ON JUNE2025 4.45 LAKHS VIEWS ON BLOG WORLDREACH AVAILABLEFOR YOUR ADVERTISEMENT
join me on Linkedin
Anthony Melvin Crasto Ph.D – India | LinkedIn
join me on Researchgate
RESEARCHGATE
join me on Facebook
Anthony Melvin Crasto Dr. | Facebook
join me on twitter
Anthony Melvin Crasto Dr. | twitter
+919321316780 call whatsaapp
EMAIL. amcrasto@gmail.com
……
| Clinical data | |
|---|---|
| Other names | UBX1325 |
| Identifiers | |
| IUPAC name | |
| CAS Number | 2271269-01-1 |
| PubChem CID | 147562879 |
| IUPHAR/BPS | 13366 |
| ChemSpider | 115277082 |
| UNII | VT53CL5GES |
| Chemical and physical data | |
| Formula | C53H59ClF3N6O10PS3 |
| Molar mass | 1159.69 g·mol−1 |
| 3D model (JSmol) | Interactive image |
| SMILES | |
| InChI | |
References
- Crago SM (22 June 2023). “Design for Phase 2B ASPIRE Study of UBX1325 for DME announced by UNITY”. Modern Retina. Archived from the original on 13 August 2024.
- Macha N, Yu M, Sapieha P, Klier S, Ghosh A, White L, et al. (September 2024). “Multifocal Electroretinography Changes after UBX1325 (Foselutoclax) Treatment in Neovascular Age-Related Macular Degeneration”. Journal of Clinical Medicine. 13 (18): 5540. doi:10.3390/jcm13185540. PMC 11433175. PMID 39337030.
//////////foselutoclax, antineoplastic, VT53CL5GES, UBX 1325
Ezobresib
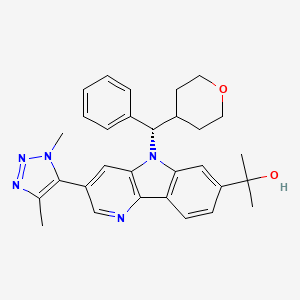
Ezobresib
CAS 1800340-40-2
MF C30H33N5O2 MW 495.6 g/mol
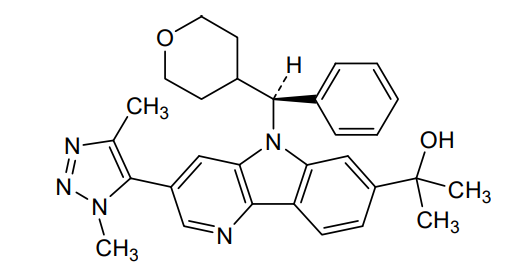
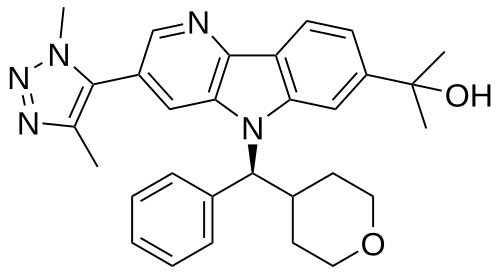
2-{3-(1,4-dimethyl-1H-1,2,3-triazol-5-yl)-5-[(S)-(oxan-4-yl)(phenyl)methyl]-5H-pyrido[3,2-b]indol-7-yl}propan-2-ol
bromodomain and extra-terminal motif (BET) inhibitor,
antineoplastic, BMS-986158, BMS 986158, Bristol Myers Squibb, antineoplastic, UNII-X8BW0MQ5PI
2-[3-(3,5-dimethyltriazol-4-yl)-5-[(S)-oxan-4-yl(phenyl)methyl]pyrido[3,2-b]indol-7-yl]propan-2-ol
Ezobresib is an investigational new drug that has been evaluated for the treatment of cancer. It inhibits Bromodomain and Extra-Terminal domain (BET) proteins, with potential antineoplastic activity.[1] Developed by Bristol Myers Squibb, this therapeutic agent has been studied for its efficacy in treating various cancers, including solid tumors and hematological malignancies.[2] Despite showing promise in early-phase clinical trials, recent developments suggest that Bristol Myers Squibb has decided to discontinue further development of ezobresib.[3]
BMS-986158 is under investigation in clinical trial NCT02419417 (Study of BMS-986158 in Subjects With Select Advanced Cancers).
Ezobresib is an inhibitor of the Bromodomain (BRD) and Extra-Terminal domain (BET) family of proteins, with potential antineoplastic activity. Upon administration, ezobresib binds to the acetyl-lysine binding site in the BRD of BET proteins, thereby preventing the interaction between BET proteins and acetylated histones. This disrupts chromatin remodeling and prevents the expression of certain growth-promoting genes, resulting in an inhibition of tumor cell growth. BET proteins (BRD2, BRD3, BRD4 and BRDT) are transcriptional regulators that bind to acetylated lysines on the tails of histones H3 and H4, and regulate chromatin structure and function; they play an important role in the modulation of gene expression during development and cellular growth
SYN
https://patentscope.wipo.int/search/en/detail.jsf?docId=US206490064&_cid=P21-MGLNPO-16484-1
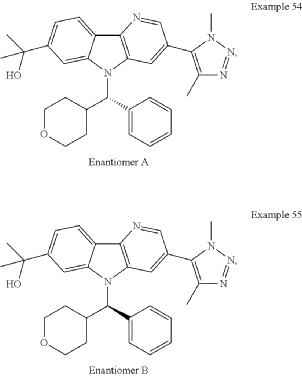
Examples 54 & 55
2-[3-(Dimethyl-1H-1,2,3-triazol-5-yl)-5-[oxan-4-yl(phenyl)methyl]-5H-pyrido[3,2-b]indol-7-yl]propan-2-ol
Step 1: 2-Chloro-5-(1,4-dimethyl-1H-1,2,3-triazol-5-yl)pyridin-3-amine
Step 2: Methyl 3-((2-chloro-5-(1,4-dimethyl-1H-1,2,3-triazol-5-yl)pyridin-3-yl)amino)benzoate
Step 3: Methyl 3-(1,4-dimethyl-1H-1,2,3-triazol-5-yl)-5H-pyrido[3,2-b]indole-7-carboxylate
Alternate synthesis of Methyl 3-(1,4-dimethyl-1H-1,2,3-triazol-5-yl)-5H-pyrido[3,2-b]indole-7-carboxylate
Step 4: Methyl 3-(1,4-dimethyl-1H-1,2,3-triazol-5-yl)-5-(phenyl(tetrahydro-2H-pyran-4-yl)methyl)-5H-pyrido[3,2-b]indole-7-carboxylate
Step 5: 2-[3-(Dimethyl-1H-1,2,3-triazol-5-yl)-5-[oxan-4-yl(phenyl)methyl]-5H-pyrido[3,2-b]indol-7-yl]propan-2-ol
Alternate Synthesis of Examples 54
2-[3-(Dimethyl-1H-1,2,3-triazol-5-yl)-5-[oxan-4-yl(phenyl)methyl]-5H-pyrido[3,2-b]indol-7-yl]propan-2-ol
Step 1: 2-Chloro-5-(1,4-dimethyl-1H-1,2,3-triazol-5-yl)pyridin-3-amine
Step 2: Methyl 3-((2-chloro-5-(1,4-dimethyl-1H-1,2,3-triazol-5-yl)pyridin-3-yl)amino)benzoate
Step 3: Methyl 3-(1,4-dimethyl-1H-1,2,3-triazol-5-yl)-5H-pyrido[3,2-b]indole-7-carboxylate
Alternate synthesis of Methyl 3-(1,4-dimethyl-1H-1,2,3-triazol-5-yl)-5H-pyrido[3,2-b]indole-7-carboxylate
Step 4: Methyl 3-(1,4-dimethyl-1H-1,2,3-triazol-5-yl)-5-(phenyl(tetrahydro-2H-pyran-4-yl)methyl)-5H-pyrido[3,2-b]indole-7-carboxylate
Step 5: 2-[3-(Dimethyl-1H-1,2,3-triazol-5-yl)-5-[oxan-4-yl(phenyl)methyl]-5H-pyrido[3,2-b]indol-7-yl]propan-2-ol
Alternate Synthesis of Examples 54
2-[3-(Dimethyl-1H-1,2,3-triazol-5-yl)-5-[oxan-4-yl(phenyl)methyl]-5H-pyrido[3,2-b]indol-7-yl]propan-2-ol
Step 1: (S)-methyl 3-(1,4-dimethyl-1H-1,2,3-triazol-5-yl)-5-(phenyl(tetrahydro-2H-pyran-4-yl)methyl)-5H-pyrido[3,2-b]indole-7-carboxylate
Step 2. (S)-2-[3-(Dimethyl-1H-1,2,3-triazol-5-yl)-5-[oxan-4-yl(phenyl)methyl]-5H-pyrido[3,2-b]indol-7-yl]propan-2-ol
Step 1: (S)-methyl 3-(1,4-dimethyl-1H-1,2,3-triazol-5-yl)-5-(phenyl(tetrahydro-2H-pyran-4-yl)methyl)-5H-pyrido[3,2-b]indole-7-carboxylate
Step 2. (S)-2-[3-(Dimethyl-1H-1,2,3-triazol-5-yl)-5-[oxan-4-yl(phenyl)methyl]-5H-pyrido[3,2-b]indol-7-yl]propan-2-ol
PATENT
LIT
- BLM overexpression as a predictive biomarker for CHK1 inhibitor response in PARP inhibitor–resistant BRCA -mutant ovarian cancerPublication Name: Science Translational MedicinePublication Date: 2023-06-21PMCID: PMC10758289PMID: 37343085DOI: 10.1126/scitranslmed.add7872
- Recent updates on 1,2,3-triazole-containing hybrids with in vivo therapeutic potential against cancers: A mini-reviewPublication Name: European Journal of Medicinal ChemistryPublication Date: 2023-05-05PMID: 36893627DOI: 10.1016/j.ejmech.2023.115254
- Development of BET Inhibitors as Potential Treatments for Cancer: Optimization of Pharmacokinetic PropertiesPublication Name: ACS Medicinal Chemistry LettersPublication Date: 2022-07-05PMCID: PMC9290009PMID: 35859878DOI: 10.1021/acsmedchemlett.2c00219
- Synthesis of BMS-986158Publication Name: SynfactsPublication Date: 2021-11-17DOI: 10.1055/s-0041-1737090
- Development of BET inhibitors as potential treatments for cancer: A new carboline chemotypePublication Name: Bioorganic & Medicinal Chemistry LettersPublication Date: 2021-11-01PMID: 34560263DOI: 10.1016/j.bmcl.2021.128376
- Discovery and Preclinical Pharmacology of an Oral Bromodomain and Extra-Terminal (BET) Inhibitor Using Scaffold-Hopping and Structure-Guided Drug DesignPublication Name: Journal of Medicinal ChemistryPublication Date: 2021-09-20PMID: 34543572DOI: 10.1021/acs.jmedchem.1c00625
- Retrospective assessment of rat liver microsomal stability at NCATS: data and QSAR modelsPublication Name: Scientific ReportsPublication Date: 2020-11-26PMCID: PMC7693334PMID: 33244000DOI: 10.1038/s41598-020-77327-0
- High-Throughput Screening to Identify Inhibitors of the Type I Interferon–Major Histocompatibility Complex Class I Pathway in Skeletal MusclePublication Name: ACS Chemical BiologyPublication Date: 2020-05-27PMCID: PMC7859889PMID: 32459468DOI: 10.1021/acschembio.0c00343
- Predictive models of aqueous solubility of organic compounds built on A large dataset of high integrityPublication Name: Bioorganic & Medicinal ChemistryPublication Date: 2019-07-15PMCID: PMC8274818PMID: 31176566DOI: 10.1016/j.bmc.2019.05.037
- Highly predictive and interpretable models for PAMPA permeabilityPublication Name: Bioorganic & Medicinal ChemistryPublication Date: 2017-02-01PMCID: PMC5291813PMID: 28082071DOI: 10.1016/j.bmc.2016.12.049
- BET inhibitor resistance emerges from leukaemia stem cellsPublication Name: NaturePublication Date: 2015-09-14PMCID: PMC6069604PMID: 26367796DOI: 10.1038/nature14888
- Efficacy of BET Bromodomain Inhibition in Kras-Mutant Non–Small Cell Lung CancerPublication Name: Clinical cancer research : an official journal of the American Association for Cancer ResearchPublication Date: 2013-11-14PMCID: PMC3838895PMID: 24045185DOI: 10.1158/1078-0432.ccr-12-3904
- Discovery and Preclinical Evaluation of [4-[[1-(3-fluorophenyl)methyl]-1H-indazol-5-ylamino]-5-methylpyrrolo[2,1-f][1,2,4]triazin-6-yl]carbamic Acid, (3S)-3-Morpholinylmethyl Ester (BMS-599626), a Selective and Orally Efficacious Inhibitor of Human Epidermal Growth Factor Receptor 1 and 2 KinasesPublication Name: Journal of Medicinal ChemistryPublication Date: 2009-10-12PMID: 19821562DOI: 10.1021/jm9010065
- [Statistical analysis of cerebrospinal fluid acid-base equilibrium and cerebrospinal fluid lactate concentration in cases of brain tumors, cerebrocranial injuries and meningoencephalitis]Publication Name: Neurologia i neurochirurgia polskaPublication Date: 1976-07PMID: 8740
PAT
- Tricyclic compounds as anticancer agentsPublication Number: WO-2015100282-A1Priority Date: 2013-12-24
- Tricyclic compound as anticancer agentsPublication Number: EP-3466949-B1Priority Date: 2013-12-24Grant Date: 2020-12-23
- Novel tricyclic compounds as anticancer agentsPublication Number: TW-202028203-APriority Date: 2013-12-24
- Novel tricyclic compounds as anticancer agentsPublication Number: TW-I726544-BPriority Date: 2013-12-24Grant Date: 2021-05-01
- Tricyclic compounds as anticancer agentsPublication Number: CN-108558871-BPriority Date: 2013-12-24Grant Date: 2022-02-18
AS ON JUNE2025 4.45 LAKHS VIEWS ON BLOG WORLDREACH AVAILABLEFOR YOUR ADVERTISEMENT
join me on Linkedin
Anthony Melvin Crasto Ph.D – India | LinkedIn
join me on Researchgate
RESEARCHGATE
join me on Facebook
Anthony Melvin Crasto Dr. | Facebook
join me on twitter
Anthony Melvin Crasto Dr. | twitter
+919321316780 call whatsaapp
EMAIL. amcrasto@gmail.com
……
| Clinical data | |
|---|---|
| Other names | BMS-986158 |
| Identifiers | |
| IUPAC name | |
| CAS Number | 1800340-40-2 |
| PubChem CID | 118196485 |
| DrugBank | DB15435 |
| ChemSpider | 58828664 |
| UNII | X8BW0MQ5PI |
| KEGG | D12710 |
| ChEMBL | ChEMBL4297458 |
| Chemical and physical data | |
| Formula | C30H33N5O2 |
| Molar mass | 495.627 g·mol−1 |
| 3D model (JSmol) | Interactive image |
| SMILES | |
| InChI | |
References
- Ma Z, Zhang C, Bolinger AA, Zhou J (October 2024). “An updated patent review of BRD4 degraders”. Expert Opinion on Therapeutic Patents. 34 (10): 929–951. doi:10.1080/13543776.2024.2400166. PMC 11427152. PMID 39219068.
- “Clinical Trials Using Ezobresib”. National Cancer Institute.
- Brown A. “Bristol backs out of BET inhibition”. ApexOnco.
////////////Ezobresib, antineoplastic, BMS-986158, BMS 986158, Bristol Myers Squibb, antineoplastic, UNII-X8BW0MQ5PI
Dapolsertib
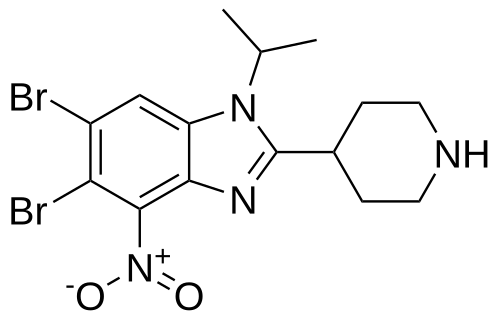
Dapolsertib
CAS 1616359-00-2
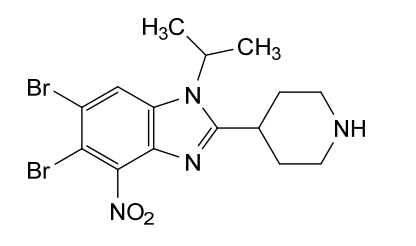
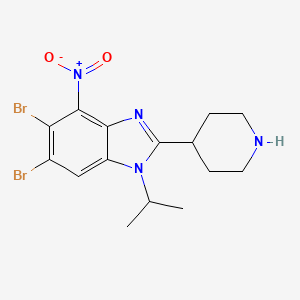
MF C15H18Br2N4O MW 446.14 g/mol
5,6-dibromo-4-nitro-2-piperidin-4-yl-1-propan-2-ylbenzimidazole
5,6-dibromo-4-nitro-2-(piperidin-4-yl)-1-(propan-2-yl)-1H-1,3-benzimidazole
serine/ threonine kinase inhibitor, antineoplastic
Ryvu Therapeutics SA, MEN1703, SEL24-B489
- SEL24-B489
- SEL-24 free base
- 9M7X64VTLI
- SEL-24
Dapolsertib is an investigational new drug that is being evaluated for the treatment of cancer. It is dual inhibitor of PIM family of serine/threonine protein kinases and mutant forms of FMS-related tyrosine kinase 3 (FLT3) that is being developed by Ryvu Therapeutics SA.[1]
Dapolsertib is an orally available inhibitor of PIM family serine/threonine protein kinases and mutant forms of FMS-related tyrosine kinase 3 (FLT3; STK1) with potential antineoplastic activity. Upon oral administration, dapolsertib binds to and inhibits the kinase activities of PIM-1, -2 and -3, and mutant forms of FLT3, which may result in the interruption of the G1/S phase cell cycle transition, an inhibition of cell proliferation, and an induction of apoptosis in tumor cells that overexpress PIMs or express mutant forms of FLT3. FLT3, a tyrosine kinase receptor that is overexpressed or mutated in various cancers, plays a role in signaling pathways that regulate hematopoietic progenitor cell proliferation, and in leukemic cell proliferation and survival. PIM kinases, downstream effectors of many cytokine and growth factor signaling pathways, including the FLT3 signaling pathway, play key roles in cell cycle progression and apoptosis inhibition and may be overexpressed in various malignancies.
- MEN1703 (SEL24) in Participants With Acute Myeloid LeukemiaCTID: NCT03008187Phase: Phase 1/Phase 2Status: CompletedDate: 2025-04-29
- MEN1703 (SEL24) to Treat Relapsed or Refractory Aggressive B-cell Non-Hodgkin Lymphoma (JASPIS-01)CTID: NCT06534437Phase: Phase 2Status: RecruitingDate: 2025-04-11
PAT
https://patentscope.wipo.int/search/en/detail.jsf?docId=WO2014096388&_cid=P12-MG5YKY-59978-1
3.9. Compounds of Example 26:
3.9. Compounds of Example 26:
5,6-dibromo-4-nitro-2-(piperidin-4-yl)-1-(propan-2-yl)-1H-1,3-benzodiazole (Example 26A):
4,5-dibromo-1-N-(propan-2-yl)benzene-1,2-diamine (2,8g, 9,lmmol) and
isonipeconic acid (1,17g, 9,lmmol) were taken up in phosphoric acid (17,82g, 0,18mol). The resulting mixture was stirred at 180°C for 3,5 hours. The mixture was allowed to cool to RT and diluted with water to 200ml. The solution was basified to pH 14.0 using solid NaOH. The resulting precipitate was then filtered off and washed repeatedly with MeOH. The filtrate was concentrated in-vacuo. The product was purified on Al2O3 (basic) using DCM/MeOH/NH3 sat. in MEOH (25: 15: 1). The obtained product (8,7mmol, 3,9g) was dissolved in cone. H2SO4 (30ml). Next KNO3 (8,7mmol, 0,89g) was added in one portion at 0° C. The resulting mixture was stirred at 0°C for 3h and at RT overnight. Then the mixture was poured onto ice. The product was filtered and washed with water.The product was purified on on Al2O3 (basic) using DCM/MeOH/NH3 sat. in MEOH (25: 15: 1) to afford 5,6-dibromo-4- nitro-2-(piperidin-4-yl)-1-(propan-2-yl)-1H-1,3-benzodiazole (1,9g). 1H NMR (600 MHz, DMSO) δ 8.74 (bs, 1H), 8.48 (s, 1H), 8.35 (bs, 1H), 4.94 (hept, J = 6.8 Hz, 1H), 3.52 – 3.46 (m, 1H), 3.42 – 3.37 (m, 2H), 3.08 (bs, 2H), 2.07 – 1.96 (m, 4H), 1.60 (d, J = 6.9 Hz, 6H). m/z 446,8; rt 2,7min.
5,6-dibromo-4-nitro-2-(piperidin-4-yl)-1-(propan-2-yl)-1H-1,3-benzodiazole (Example 26A):
4,5-dibromo-1-N-(propan-2-yl)benzene-1,2-diamine (2,8g, 9,lmmol) and
isonipeconic acid (1,17g, 9,lmmol) were taken up in phosphoric acid (17,82g, 0,18mol). The resulting mixture was stirred at 180°C for 3,5 hours. The mixture was allowed to cool to RT and diluted with water to 200ml. The solution was basified to pH 14.0 using solid NaOH. The resulting precipitate was then filtered off and washed repeatedly with MeOH. The filtrate was concentrated in-vacuo. The product was purified on Al2O3 (basic) using DCM/MeOH/NH3 sat. in MEOH (25: 15: 1). The obtained product (8,7mmol, 3,9g) was dissolved in cone. H2SO4 (30ml). Next KNO3 (8,7mmol, 0,89g) was added in one portion at 0° C. The resulting mixture was stirred at 0°C for 3h and at RT overnight. Then the mixture was poured onto ice. The product was filtered and washed with water.The product was purified on on Al2O3 (basic) using DCM/MeOH/NH3 sat. in MEOH (25: 15: 1) to afford 5,6-dibromo-4- nitro-2-(piperidin-4-yl)-1-(propan-2-yl)-1H-1,3-benzodiazole (1,9g). 1H NMR (600 MHz, DMSO) δ 8.74 (bs, 1H), 8.48 (s, 1H), 8.35 (bs, 1H), 4.94 (hept, J = 6.8 Hz, 1H), 3.52 – 3.46 (m, 1H), 3.42 – 3.37 (m, 2H), 3.08 (bs, 2H), 2.07 – 1.96 (m, 4H), 1.60 (d, J = 6.9 Hz, 6H). m/z 446,8; rt 2,7min.
PAT
Novel benzimidazole derivatives as kinase inhibitors
Publication Number: WO-2014096388-A2
Priority Date: 2012-12-21
- Novel benzimidazole derivatives as kinase inhibitorsPublication Number: KR-20150095908-APriority Date: 2012-12-21
- Benzimidazole derivatives as kinase inhibitorsPublication Number: US-10174013-B2Priority Date: 2012-12-21Grant Date: 2019-01-08
- Novel Benzimidazole Derivatives as Kinase InhibitorsPublication Number: US-2015336967-A1Priority Date: 2012-12-21
- Novel benzimidazole derivatives as kinase inhibitorsPublication Number: US-2017152249-A1Priority Date: 2012-12-21
- Benzimidazole derivatives as Kinase InhibitorsPublication Number: US-9388192-B2Priority Date: 2012-12-21Grant Date: 2016-07-12
- Novel benzimidazole derivatives as kinase inhibitorsPublication Number: EP-2935244-B1Priority Date: 2012-12-21Grant Date: 2018-06-27
- Novel benzimidazole derivatives as kinase inhibitorsPublication Number: ES-2688395-T3Priority Date: 2012-12-21Grant Date: 2018-11-02
- Novel benzimidazole derivatives as kinase inhibitorsPublication Number: JP-2016503779-APriority Date: 2012-12-21
- Novel benzimidazole derivatives as kinase inhibitorsPublication Number: JP-6169185-B2Priority Date: 2012-12-21Grant Date: 2017-07-26
- Novel benzimidazole derivatives as kinase inhibitorsPublication Number: KR-101779272-B1Priority Date: 2012-12-21Grant Date: 2017-09-18
AS ON JUNE2025 4.45 LAKHS VIEWS ON BLOG WORLDREACH AVAILABLEFOR YOUR ADVERTISEMENT
join me on Linkedin
Anthony Melvin Crasto Ph.D – India | LinkedIn
join me on Researchgate
RESEARCHGATE
join me on Facebook
Anthony Melvin Crasto Dr. | Facebook
join me on twitter
Anthony Melvin Crasto Dr. | twitter
+919321316780 call whatsaapp
EMAIL. amcrasto@gmail.com
……
| Clinical data | |
|---|---|
| Other names | MEN1703, SEL24-B489 |
| Identifiers | |
| IUPAC name | |
| CAS Number | 1616359-00-2 |
| PubChem CID | 76286825 |
| IUPHAR/BPS | 13204 |
| ChemSpider | 81367232 |
| UNII | 9M7X64VTLI |
| ChEMBL | ChEMBL4467168 |
| Chemical and physical data | |
| Formula | C15H18Br2N4O2 |
| Molar mass | 446.143 g·mol−1 |
| 3D model (JSmol) | Interactive image |
| SMILES | |
| InChI | |
References
- Wu M, Li C, Zhu X (December 2018). “FLT3 inhibitors in acute myeloid leukemia”. Journal of Hematology & Oncology. 11 (1) 133. doi:10.1186/s13045-018-0675-4. PMC 6280371. PMID 30514344.
//////////Dapolsertib, antineoplastic, MEN1703, SEL24-B489, MEN 1703, SEL24 B489, Ryvu Therapeutics SA
Ropeginterferon alfa-2b
PCDLPQTHSL GSRRTLMLLA QMRRISLFSC LKDRHDFGFP QEEFGNQFQK AETIPVLHEM
IQQIFNLFST KDSSAAWDET LLDKFYTELY QQLNDLEACV IQGVGVTETP LMKEDSILAV
RKYFQRITLY LKEKKYSPCA WEVVRAEIMR SFSLSTNLQE SLRSKE
(Disulfide bridge: 2-99, 30-139)
Ropeginterferon alfa-2b
- AOP2014
CAS 1335098-50-4
UNII981TME683S
FDA APPROVED, 2021/11/12, BESREMI
PEPTIDE, Antineoplastic, Antiviral
Polycythemia vera (PV) is the most common Philadelphia chromosome-negative myeloproliferative neoplasm (MPN), characterized by increased hematocrit and platelet/leukocyte counts, an increased risk for hemorrhage and thromboembolic events, and a long-term propensity for myelofibrosis and leukemia.1,2 Interferon alfa-2b has been used for decades to treat PV but requires frequent dosing and is not tolerated by all patients.2 Ropeginterferon alfa-2b is a next-generation mono-pegylated type I interferon produced from proline-IFN-α-2b in Escherichia coli that has high tolerability and a long half-life.4,6 Ropeginterferon alfa-2b has shown efficacy in PV in in vitro and in vivo models and clinical trials.3,4
Ropeginterferon alfa-2b was approved by the FDA on November 12, 2021, and is currently marketed under the trademark BESREMi by PharmaEssentia Corporation.6
Ropeginterferon alfa-2b, sold under the brand name Besremi, is a medication used to treat polycythemia vera.[1][2][3][4] It is an interferon.[1][3] It is given by injection.[1][3]
The most common side effects include low levels of white blood cells and platelets (blood components that help the blood to clot), muscle and joint pain, tiredness, flu-like symptoms and increased blood levels of gamma-glutamyl transferase (a sign of liver problems).[3] Ropeginterferon alfa-2b can cause liver enzyme elevations, low levels of white blood cells, low levels of platelets, joint pain, fatigue, itching, upper airway infection, muscle pain and flu-like illness.[2] Side effects may also include urinary tract infection, depression and transient ischemic attacks (stroke-like attacks).[2]
It was approved for medical use in the European Union in February 2019,[3] and in the United States in November 2021.[2][5] Ropeginterferon alfa-2b is the first medication approved by the U.S. Food and Drug Administration (FDA) to treat polycythemia vera that people can take regardless of their treatment history, and the first interferon therapy specifically approved for polycythemia vera.[2]
https://www.fda.gov/news-events/press-announcements/fda-approves-treatment-rare-blood-disease#:~:text=FDA%20NEWS%20RELEASE-,FDA%20Approves%20Treatment%20for%20Rare%20Blood%20Disease,FDA%2DApproved%20Option%20Patients%20Can%20Take%20Regardless%20of%20Previous%20Therapies,-ShareFor Immediate Release:November 12, 2021
Today, the U.S. Food and Drug Administration approved Besremi (ropeginterferon alfa-2b-njft) injection to treat adults with polycythemia vera, a blood disease that causes the overproduction of red blood cells. The excess cells thicken the blood, slowing blood flow and increasing the chance of blood clots.
“Over 7,000 rare diseases affect more than 30 million people in the United States. Polycythemia vera affects approximately 6,200 Americans each year,” said Ann Farrell, M.D., director of the Division of Non-Malignant Hematology in the FDA’s Center for Drug Evaluation and Research. “This action highlights the FDA’s commitment to helping make new treatments available to patients with rare diseases.”
Besremi is the first FDA-approved medication for polycythemia vera that patients can take regardless of their treatment history, and the first interferon therapy specifically approved for polycythemia vera.
Treatment for polycythemia vera includes phlebotomies (a procedure that removes excess blood cells though a needle in a vein) as well as medicines to reduce the number of blood cells; Besremi is one of these medicines. Besremi is believed to work by attaching to certain receptors in the body, setting off a chain reaction that makes the bone marrow reduce blood cell production. Besremi is a long-acting drug that patients take by injection under the skin once every two weeks. If Besremi can reduce excess blood cells and maintain normal levels for at least one year, then dosing frequency may be reduced to once every four weeks.
The effectiveness and safety of Besremi were evaluated in a multicenter, single-arm trial that lasted 7.5 years. In this trial, 51 adults with polycythemia vera received Besremi for an average of about five years. Besremi’s effectiveness was assessed by looking at how many patients achieved complete hematological response, which meant that patients had a red blood cell volume of less than 45% without a recent phlebotomy, normal white cell counts and platelet counts, a normal spleen size, and no blood clots. Overall, 61% of patients had a complete hematological response.
Besremi can cause liver enzyme elevations, low levels of white blood cells, low levels of platelets, joint pain, fatigue, itching, upper airway infection, muscle pain and flu-like illness. Side effects may also include urinary tract infection, depression and transient ischemic attacks (stroke-like attacks).
Interferon alfa products like Besremi may cause or worsen neuropsychiatric, autoimmune, ischemic (not enough blood flow to a part of the body) and infectious diseases, which could lead to life-threatening or fatal complications. Patients who must not take Besremi include those who are allergic to the drug, those with a severe psychiatric disorder or a history of a severe psychiatric disorder, immunosuppressed transplant recipients, certain patients with autoimmune disease or a history of autoimmune disease, and patients with liver disease.
People who could be pregnant should be tested for pregnancy before using Besremi due to the risk of fetal harm.
Besremi received orphan drug designation for this indication. Orphan drug designation provides incentives to assist and encourage drug development for rare diseases.
The FDA granted the approval of Besremi to PharmaEssentia Corporation.
Medical uses
In the European Union, ropeginterferon alfa-2b is indicated as monotherapy in adults for the treatment of polycythemia vera without symptomatic splenomegaly.[3] In the United States it is indicated for the treatment of polycythemia vera.[1][2][5]
History
The effectiveness and safety of ropeginterferon alfa-2b were evaluated in a multicenter, single-arm trial that lasted 7.5 years.[2] In this trial, 51 adults with polycythemia vera received ropeginterferon alfa-2b for an average of about five years.[2] The effectiveness of ropeginterferon alfa-2b was assessed by looking at how many participants achieved complete hematological response, which meant that participants had a red blood cell volume of less than 45% without a recent phlebotomy, normal white cell counts and platelet counts, a normal spleen size, and no blood clots.[2] Overall, 61% of participants had a complete hematological response.[2] The U.S. Food and Drug Administration (FDA) granted the application for Ropeginterferon_alfa-2b orphan drug designation and granted the approval of Besremi to PharmaEssentia Corporation[2]
REF
- Bartalucci N, Guglielmelli P, Vannucchi AM: Polycythemia vera: the current status of preclinical models and therapeutic targets. Expert Opin Ther Targets. 2020 Jul;24(7):615-628. doi: 10.1080/14728222.2020.1762176. Epub 2020 May 18. [Article]
- How J, Hobbs G: Use of Interferon Alfa in the Treatment of Myeloproliferative Neoplasms: Perspectives and Review of the Literature. Cancers (Basel). 2020 Jul 18;12(7). pii: cancers12071954. doi: 10.3390/cancers12071954. [Article]
- Verger E, Soret-Dulphy J, Maslah N, Roy L, Rey J, Ghrieb Z, Kralovics R, Gisslinger H, Grohmann-Izay B, Klade C, Chomienne C, Giraudier S, Cassinat B, Kiladjian JJ: Ropeginterferon alpha-2b targets JAK2V617F-positive polycythemia vera cells in vitro and in vivo. Blood Cancer J. 2018 Oct 4;8(10):94. doi: 10.1038/s41408-018-0133-0. [Article]
- Gisslinger H, Zagrijtschuk O, Buxhofer-Ausch V, Thaler J, Schloegl E, Gastl GA, Wolf D, Kralovics R, Gisslinger B, Strecker K, Egle A, Melchardt T, Burgstaller S, Willenbacher E, Schalling M, Them NC, Kadlecova P, Klade C, Greil R: Ropeginterferon alfa-2b, a novel IFNalpha-2b, induces high response rates with low toxicity in patients with polycythemia vera. Blood. 2015 Oct 8;126(15):1762-9. doi: 10.1182/blood-2015-04-637280. Epub 2015 Aug 10. [Article]
- EMA Approved Products: Besremi (ropeginterferon alfa-2b ) solution for injection [Link]
- FDA Approved Drug Products: BESREMi (ropeginterferon alfa-2b-njft) injection [Link]
join me on Linkedin
Anthony Melvin Crasto Ph.D – India | LinkedIn
join me on Researchgate
RESEARCHGATE
join me on Facebook
Anthony Melvin Crasto Dr. | Facebook
join me on twitter
Anthony Melvin Crasto Dr. | twitter
+919321316780 call whatsaapp
EMAIL. amcrasto@amcrasto
/////////////////////////////////////////////////////////////////////////////////////////////////////
References
- ^ Jump up to:a b c d e https://www.accessdata.fda.gov/drugsatfda_docs/label/2021/761166s000lbl.pdf
- ^ Jump up to:a b c d e f g h i j k l “FDA Approves Treatment for Rare Blood Disease”. U.S. Food and Drug Administration (FDA) (Press release). 12 November 2021. Retrieved 12 November 2021.
 This article incorporates text from this source, which is in the public domain.
- ^ Jump up to:a b c d e f g “Besremi EPAR”. European Medicines Agency (EMA). Retrieved 14 November 2021. Text was copied from this source which is copyright European Medicines Agency. Reproduction is authorized provided the source is acknowledged.
- ^ Wagner SM, Melchardt T, Greil R (March 2020). “Ropeginterferon alfa-2b for the treatment of patients with polycythemia vera”. Drugs of Today. Barcelona, Spain. 56 (3): 195–202. doi:10.1358/dot.2020.56.3.3107706. PMID 32282866. S2CID 215758794.
- ^ Jump up to:a b “U.S. FDA Approves Besremi (ropeginterferon alfa-2b-njft) as the Only Interferon for Adults With Polycythemia Vera” (Press release). PharmaEssentia. 12 November 2021. Retrieved 14 November 2021 – via Business Wire.
External links
- “Ropeginterferon alfa-2b”. Drug Information Portal. U.S. National Library of Medicine.
- Clinical trial number NCT01193699 for “Safety Study of Pegylated Interferon Alpha 2b to Treat Polycythemia Vera (PEGINVERA)” at ClinicalTrials.gov
- Clinical trial number NCT02218047 for “AOP2014 vs. BAT in Patients With Polycythemia Vera Who Previously Participated in the PROUD-PV Study. (CONTI-PV)” at ClinicalTrials.gov
| Clinical data | |
|---|---|
| Trade names | Besremi |
| Other names | AOP2014, ropeginterferon alfa-2b-njft |
| License data | EU EMA: by INNUS DailyMed: Ropeginterferon_alfa |
| Pregnancy category | Contraindicated |
| Routes of administration | Subcutaneous |
| Drug class | Interferon |
| ATC code | L03AB15 (WHO) |
| Legal status | |
| Legal status | US: ℞-only [1][2]EU: Rx-only [3] |
| Identifiers | |
| CAS Number | 1335098-50-4 |
| DrugBank | DB15119 |
| UNII | 981TME683S |
| KEGG | D11027 |
/////////Ropeginterferon alfa-2b, FDA 2021, APPROVALS 2021, BESREMI, PEPTIDE, Antineoplastic, Antiviral, AOP 2014, PharmaEssentia
NEW DRUG APPROVALS
ONE TIME
$10.00
Tisotumab vedotin

Tisotumab vedotin
チソツマブベドチン (遺伝子組換え)Immunoglobulin G1, anti-(human blood-coagulation factor III) (human monoclonal HuMax-TF heavy chain), disulfide with human monoclonal HuMax-TF κ-chain, dimer, tetrakis(thioether) with N-[[[4-[[N-[6-(3-mercapto-2,5-dioxo-1-pyrrolidinyl)-1-oxohexyl]-L-valyl-N5-(aminocarbonyl)-L-ornithyl]amino]phenyl]methoxy]carbonyl]-N-methyl-L-valyl-N-[(1S,2R)-4-[(2S)-2-[(1R,2R)-3-[[(1R,2S)-2-hydroxy-1-methyl-2-phenylethyl]amino]-1-methoxy-2-methyl-3-oxopropyl]-1-pyrrolidinyl]-2-methoxy-1-[(1S)-1-methylpropyl]-4-oxobutyl]-N-methyl-L-valinamide
- HuMax-TF-ADC
- Immunoglobulin G1, anti-(human tissue factor) (human monoclonal HuMax-TF heavy chain), disulfide with human monoclonal HuMax-TF κ-chain, dimer, tetrakis(thioether) with N-[[[4-[[N-[6-(3-mercapto-2,5-dioxo-1-pyrrolidinyl)-1-oxohexyl]-L-valyl-N5-(aminocarbonyl)-L-ornithyl]amino]phenyl]methoxy]carbonyl]-N-methyl-L-valyl-N-[(1S,2R)-4-[(2S)-2-[(1R,2R)-3-[[(1R,2S)-2-hydroxy-1-methyl-2-phenylethyl]amino]-1-methoxy-2-methyl-3-oxopropyl]-1-pyrrolidinyl]-2-methoxy-1-[(1S)-1-methylpropyl]-4-oxobutyl]-N-methyl-L-valinamide
Protein Sequence
Sequence Length: 1324, 448, 448, 214, 214multichain; modified (modifications unspecified)
| Formula | C6418H9906N1710O2022S44.(C68H106N11O15)n |
|---|---|
| Efficacy | Antineoplastic |
| Disease | Cervical cancer |
| Comment | Antibody-drug conjugateCAS:1418731-10-8 |
- HuMax-TF-ADC
- Tisotumab vedotin
- Tisotumab vedotin [WHO-DD]
- UNII-T41737F88A
- WHO 10148
US FDA APPROVED 2021/9/20 , TIVDAK

FDA grants accelerated approval to tisotumab vedotin-tftv for recurrent or metastatic cervical cancer……….. https://www.fda.gov/drugs/resources-information-approved-drugs/fda-grants-accelerated-approval-tisotumab-vedotin-tftv-recurrent-or-metastatic-cervical-cancer
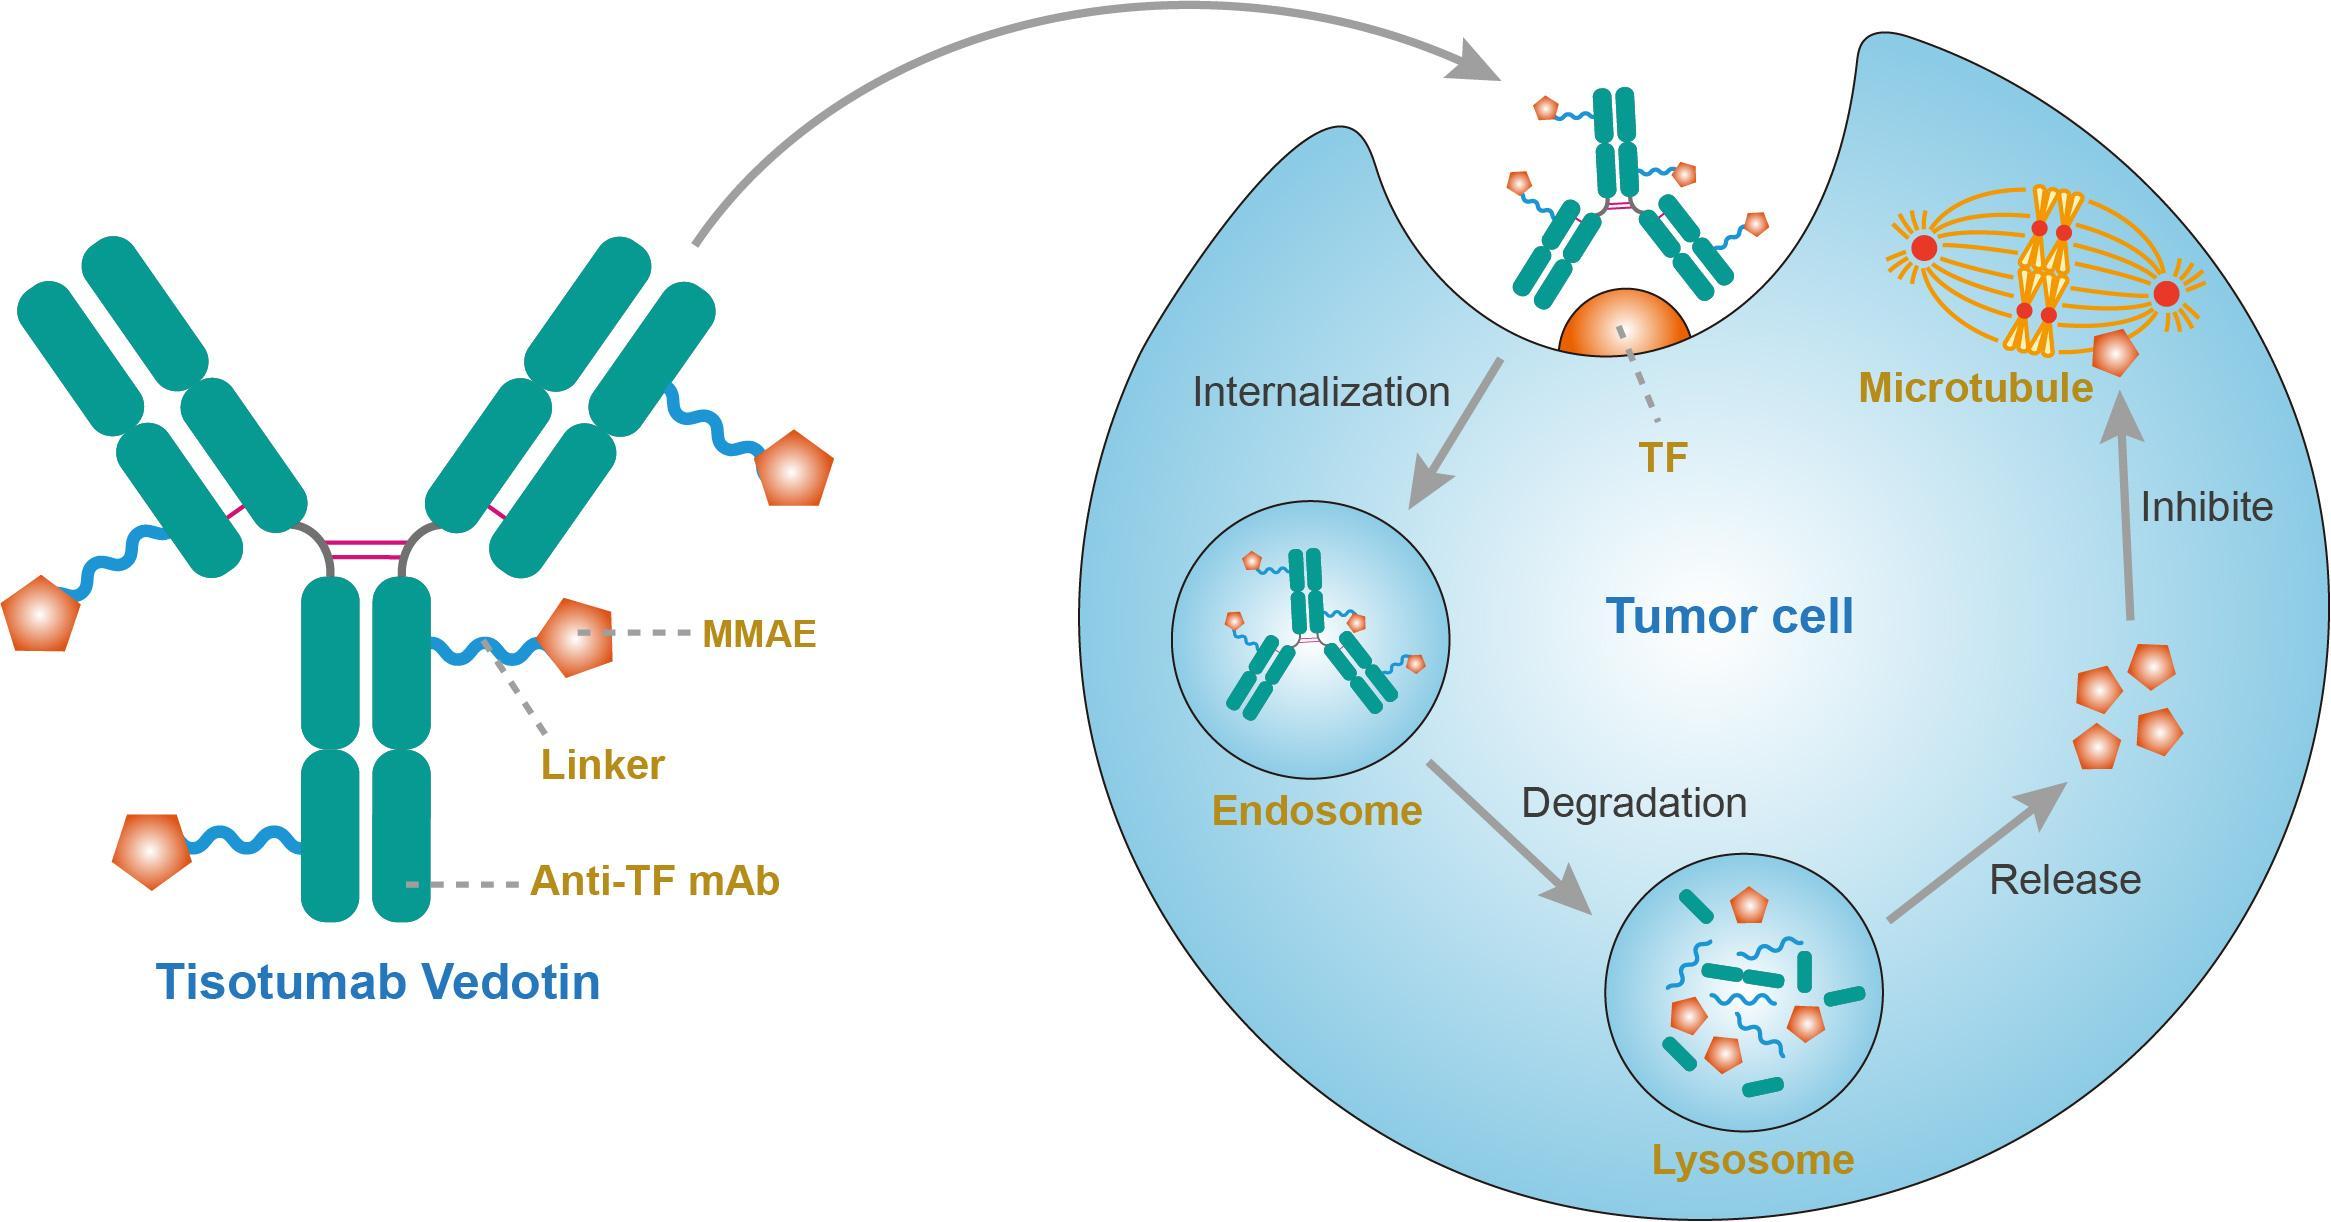
On September 20, 2021, the Food and Drug Administration granted accelerated approval to tisotumab vedotin-tftv (Tivdak, Seagen Inc.), a tissue factor-directed antibody and microtubule inhibitor conjugate, for adult patients with recurrent or metastatic cervical cancer with disease progression on or after chemotherapy.
Approval was based on innovaTV 204, an open-label, multicenter, single-arm clinical trial (NCT03438396). Efficacy was evaluated in 101 patients with recurrent or metastatic cervical cancer who had received no more than two prior systemic regimens in the recurrent or metastatic setting, including at least one prior platinum-based chemotherapy regimen. Sixty-nine percent of patients had received bevacizumab as part of prior systemic therapy. Patients received tisotumab vedotin-tftv 2 mg/kg every 3 weeks until disease progression or unacceptable toxicity.
The main efficacy outcome measures were confirmed objective response rate (ORR) as assessed by an independent review committee (IRC) using RECIST v1.1 and duration of response (DOR). The ORR was 24% (95% CI: 15.9%, 33.3%) with a median response duration of 8.3 months (95% CI: 4.2, not reached).
The most common adverse reactions (≥25%), including laboratory abnormalities, were hemoglobin decreased, fatigue, lymphocytes decreased, nausea, peripheral neuropathy, alopecia, epistaxis, conjunctival adverse reactions, hemorrhage, leukocytes decreased, creatinine increased, dry eye, prothrombin international normalized ratio increased, activated partial thromboplastin time prolonged, diarrhea, and rash. Product labeling includes a boxed warning for ocular toxicity.
The recommended dose is 2 mg/kg (up to a maximum of 200 mg for patients ≥100 kg) given as an intravenous infusion over 30 minutes every 3 weeks until disease progression or unacceptable toxicity.
View full prescribing information for Tivdak.
This review used the Assessment Aid, a voluntary submission from the applicant to facilitate the FDA’s assessment.
This application was granted priority review. A description of FDA expedited programs is in the Guidance for Industry: Expedited Programs for Serious Conditions-Drugs and Biologics.
A fully human monoclonal antibody specific for tissue factor conjugated to the microtubule-disrupting agent monomethyl auristatin E (MMAE) via a protease-cleavable valine-citrulline linker.
Tisotumab vedotin, sold under the brand name Tivdak is a human monoclonal antibody used to treat cervical cancer.[1]
Tisotumab vedotin was approved for medical use in the United States in September 2021.[1][2]
Tisotumab vedotin is the international nonproprietary name (INN).[3]
join me on Linkedin
Anthony Melvin Crasto Ph.D – India | LinkedIn
join me on Researchgate
RESEARCHGATE

join me on Facebook
Anthony Melvin Crasto Dr. | Facebook
join me on twitter
Anthony Melvin Crasto Dr. | twitter
+919321316780 call whatsaapp
EMAIL. amcrasto@amcrasto
NEW DRUG APPROVALS
one time
$10.00
References
- ^ Jump up to:a b c d https://www.accessdata.fda.gov/drugsatfda_docs/label/2021/761208s000lbl.pdf
- ^ “Seagen and Genmab Announce FDA Accelerated Approval for Tivdak (tisotumab vedotin-tftv) in Previously Treated Recurrent or Metastatic Cervical Cancer”. Seagen. 20 September 2021. Retrieved 20 September 2021 – via Business Wire.
- ^ World Health Organization (2016). “International nonproprietary names for pharmaceutical substances (INN): recommended INN: list 75”. WHO Drug Information. 30 (1): 159–60. hdl:10665/331046.
External links
- “Tisotumab vedotin”. Drug Information Portal. U.S. National Library of Medicine.
- Clinical trial number NCT03438396 for “A Trial of Tisotumab Vedotin in Cervical Cancer” at ClinicalTrials.gov
- Clinical trial number NCT03245736 for “Tisotumab Vedotin Continued Treatment in Patients With Solid Tumors” at ClinicalTrials.gov
- Clinical trial number NCT02001623 for “Tisotumab Vedotin (HuMax-TF-ADC) Safety Study in Patients With Solid Tumors” at ClinicalTrials.gov
- Clinical trial number NCT02552121 for “Tisotumab Vedotin (HuMax-TF-ADC) Safety Study in Patients With Solid Tumors” at ClinicalTrials.gov
| Monoclonal antibody | |
|---|---|
| Type | Whole antibody |
| Source | Human |
| Target | Tissue factor (TF) |
| Clinical data | |
| Trade names | Tivdak |
| Other names | Tisotumab vedotin-tftv |
| License data | US DailyMed: Tisotumab_vedotin |
| Pregnancy category | Contraindicated[1] |
| Routes of administration | Intravenous |
| Drug class | Antineoplastic |
| ATC code | None |
| Legal status | |
| Legal status | US: ℞-only [1] |
| Identifiers | |
| CAS Number | 1418731-10-8 |
| UNII | T41737F88A |
| KEGG | D11814 |
//////////Tisotumab vedotin, チソツマブベドチン (遺伝子組換え) , FDA 2021, APPROVALS 2021, Antineoplastic, CERVICAL CANCER, CANCER, MONOCLONAL ANTIBODY, UNII-T41737F88A, WHO 10148


