FLAGS AND HITS
 DRUG APPROVALS BY DR ANTHONY MELVIN CRASTO
 FACEBOOK
...................................................................Join me on twitter
 ..................................................................Join me on google plus
 Googleplus
MYSELF


TASIMELTION…FDA Approves Hetlioz: First Treatment for Non-24 Hour Sleep-Wake Disorder in Blind Individuals

TASIMELTION, an orphan drug for non24
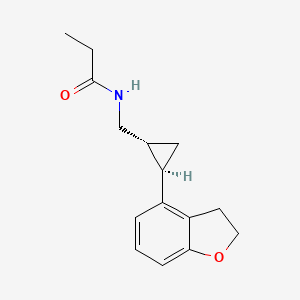
N-([(1R,2R)-2-(2,3-Dihydro-1-benzofuran-4-yl)cyclopropyl]methyl)propanamide
(1R-trans)-N-[[2-(2,3-dihydro-4-benzofuranyl)cyclopropyl]methyl]pro- pananamide VEC162
(-)-(trans)-N-[[2-(2,3-Dihydrobenzofuran-4-yl)cycloprop-1-yl]methyl]propanamide
N-(((1R,2R)-2-(2,3-Dihydro-1-benzofuran-4-yl)cyclopropyl)methyl)propanamide
Bristol-Myers Squibb Company
PRODUCT PATENT
U.S. Pat. No. 5,856,529
| CAS number | 609799-22-6 |
|---|
| Formula | C15H19NO2 |
|---|---|
| Mol. mass | 245.3 g/mol |
VEC-162, BMS-214778, 609799-22-6, Hetlioz, UNII-SHS4PU80D9,

January 31, 2014 — The U.S. Food and Drug Administration today approved Hetlioz (tasimelteon), a melatonin receptor agonist, to treat non-24- hour sleep-wake disorder (“non-24”) in totally blind individuals. Non-24 is a chronic circadian rhythm (body clock) disorder in the blind that causes problems with the timing of sleep. This is the first FDA approval of a treatment for the disorder.
Non-24 occurs in persons who are completely blind. Light does not enter their eyes and they cannot synchronize their body clock to the 24-hour light-dark cycle.
Tasimelteon
A year-long (2011-2012) study at Harvard is testing the use of tasimelteon in blind subjects with non-24-hour sleep–wake disorder.[4] In May 2013Vanda Pharmaceuticals submitted a New Drug Application to the Food and Drug Administration for Tasimelteon for the treatment of non-24-hour sleep–wake disorder in totally blind people.[5]
SEQUENCE
Discovered by Bristol-Myers Squibb (BMS) and co-developed with Vanda Pharmaceuticals, tasimelteon is a hypnotic family benzofuran. In Phase III development, it has an orphan drug status.
JAN2014.. APPROVED FDA
In mid-November 2013 the FDA announced their recommendation for the approval of Tasimelteon for the treatment of non-24-disorder.Tasimelteon effectively resets the circadian rhythm, helping to restore normal sleep patterns.http://www.fda.gov/downloads/AdvisoryCommittees/CommitteesMeetingMaterials/Drugs/PeripheralandCentralNervousSystemDrugsAdvisoryCommittee/UCM374388.pdf
January 2010: FDA granted orphan drug tasimelteon to disturbed sleep / wake in blind without light perception.
February 2008: Vanda has completed enrollment in its Phase III trial in chronic primary insomnia.
June 2007: Results of a Phase III trial for transient insomnia tasimelteon presented by Vanda at the 21st annual meeting of the Associated Professional Sleep Societies. These results demonstrated improvements in objective and subjective measures of sleep and its maintenance.
2004 Vanda gets a license tasimelteon (or BMS-214778 and VEC-162) from Bristol-Myers Squibb.
About Tasimelteon: Tasimelteon is a circadian regulator in development for the treatment of Non-24. Tasimelteon is a dual melatonin receptor agonist (DMRA) with selective agonist activityat the MT1 and MT2 receptors.Tasimelteon’s ability to reset the master body clock in the suprachiasmatic nucleus (SCN) results in the entrainment of the body’s melatonin and cortisol rhythms with the 24-hour day-night cycle. The patent claiming tasimelteon as a new chemical entity extends through December 2022, assuming a 5-year extension to be granted under the Hatch-Waxman Act. Tasimelteon has been granted orphan drug designation for the treatment of Non-24 from both the U.S. and the European Union.
Previously, BMS-214778, identified as an agonist of melatonin receptors, has been the subject of pre-clinical studies for the treatment of sleep disorders resulting from a disturbance of circadian rhythms.The first Pharmacokinetic studies were performed in rats and monkeys.
The master body clock controls the timing of many aspects of physiology, behavior and metabolism that show daily rhythms, including the sleep-wake cycles, body temperature, alertness and performance, metabolic rhythms and certain hormones which exhibit circadian variation. Outputs from the suprachiasmatic nucleus (SCN) control many endocrine rhythms including those of melatonin secretion by the pineal gland as well as the control of cortisol secretion via effects on the hypothalamus, the pituitary and the adrenal glands.
This master body clock, located in the SCN, spontaneously generates rhythms of approximately 24.5 hours. These non-24-hour rhythms are synchronized each day to the 24-hour day-night cycle by light, the primary environmental time cue which is detected by specialized cells in the retina and transmitted to the SCN via the retino-hypothalamic tract. Inability to detect this light signal, as occurs in most totally blind individuals, leads to the inability of the master body clock to be reset daily and maintain entrainment to a 24-hour day.
Non-24-Hour Disorder
Non-24, also referred to as Non-24-Hour Sleep-Wake Disorder (N24HSWD) or Non-24-Hour Disorder, is an orphan indication affecting approximately 65,000 to 95,000 people in the U.S. and 140,000 in Europe. Non-24 occurs when individuals, primarily blind with no light perception, are unable to synchronize their endogenous circadian pacemaker to the 24-hour light/dark cycle. Without light as a synchronizer, and because the period of the internal clock is typically a little longer than 24 hours, individuals with Non-24 experience their circadian drive to initiate sleep drifting later and later each day. Individuals with Non-24 have abnormal night sleep patterns, accompanied by difficulty staying awake during the day. Non-24 leads to significant impairment, with chronic effects impacting the social and occupational functioning of these individuals.
In addition to problems sleeping at the desired time, individuals with Non-24 experience excessive daytime sleepiness that often results in daytime napping. TASIMELTION
The severity of nighttime sleep complaints and/or daytime sleepiness complaints varies depending on where in the cycle the individual’s body clock is with respect to their social, work, or sleep schedule. The “free running” of the clock results in approximately a 1-4 month repeating cycle, the circadian cycle, where the circadian drive to initiate sleep continually shifts a little each day (about 15 minutes on average) until the cycle repeats itself. Initially, when the circadian cycle becomes desynchronous with the 24 h day-night cycle, individuals with Non-24 have difficulty initiating sleep. As time progresses, the internal circadian rhythms of these individuals becomes 180 degrees out of synchrony with the 24 h day-night cycle, which gradually makes sleeping at night virtually impossible, and leads to extreme sleepiness during daytime hours.
Eventually, the individual’s sleep-wake cycle becomes aligned with the night, and “free-running” individuals are able to sleep well during a conventional or socially acceptable time. However, the alignment between the internal circadian rhythm and the 24-hour day-night cycle is only temporary. In addition to cyclical nighttime sleep and daytime sleepiness problems, this condition can cause deleterious daily shifts in body temperature and hormone secretion, may cause metabolic disruption and is sometimes associated with depressive symptoms and mood disorders.
It is estimated that 50-75% of totally blind people in the United States (approximately 65,000 to 95,000) have Non-24. This condition can also affect sighted people. However, cases are rarely reported in this population, and the true rate of Non-24 in the general population is not known.
The ultimate treatment goal for individuals with Non-24 is to entrain or synchronize their circadian rhythms into an appropriate phase relationship with the 24-hour day so that they will have increased sleepiness during the night and increased wakefulness during the daytime.
INTRODUCTION
Tasimelteon has the chemical name: trans-N-[[2-(2,3-dihydrobenzofuran-4-yl)cycloprop-1yl]methyl]propanamide, has the structure of Formula I:

and is disclosed in U.S. Pat. No. 5,856,529 and in US 20090105333, both of which are incorporated herein by reference as though fully set forth.
Tasimelteon is a white to off-white powder with a melting point of about 78° C. (DSC) and is very soluble or freely soluble in 95% ethanol, methanol, acetonitrile, ethyl acetate, isopropanol, polyethylene glycols (PEG-300 and PEG-400), and only slightly soluble in water. The native pH of a saturated solution of tasimelteon in water is 8.5 and its aqueous solubility is practically unaffected by pH. Tasimelteon has 2-4 times greater affinity for MT2R relative to MT1R. It’s affinity (Ki) for MT1R is 0.3 to 0.4 and for MT2R, 0.1 to 0.2. Tasimelteon is useful in the practice of this invention because it is a melatonin agonist that has been demonstrated, among other activities, to entrain patients suffering from Non-24.
………………………..
SYNTHESIS
(1R-trans)-N-[[2 – (2,3-dihydro-4 benzofuranyl) cyclopropyl] methyl] propanamide PATENT: BRISTOL-MYERS SQUIBB PRIORITY DATE: 1996 HYPNOTIC

PREPARATION OF XV
XXIV D-camphorsulfonic acid IS REACTED WITH THIONYL CHLORIDE TO GIVE
…………XXV (1S, 4R) -7,7-dimethyl-2-oxo-bicyclo [2.2.1] heptane-1-methanesulfonyl chloride
TREATED WITH
XXVI ammonium hydroxide
TO GIVE
XXVII (1S, 4R) -7,7-dimethyl-2-oxo-bicyclo [2.2.1] heptane-1-methanesulfonamide
TREATED WITH AMBERLYST15
….XXVIII (3aS, 6R) -4,5,6,7-tetrahydro-8 ,8-dimethyl-3H-3a ,6-methano-2 ,1-benzisothiazole-2 ,2-dioxide
TREATED WITH LAH, ie double bond is reduced to get
…..XV (3aS, 6R, 7aR)-hexahydro-8 ,8-dimethyl-3H-3a ,6-methano-2 ,1-benzisothiazole-2 ,2-dioxide

Intermediate
I 3-hydroxybenzoic acid methyl ester
II 3-bromo-1-propene
III 3 – (2-propenyloxy) benzoic acid methyl ester
IV 3-hydroxy-2-(2-propenyl) benzoic acid methyl ester
V 2,3-dihydro-4-hydroxy-2-benzofurancarboxylic acid methyl ester
VI benzofuran-4-carboxylic acid methyl ester
VII benzofuran-4-carboxylic acid
VIII 2,3-dihydro-4-benzofurancarboxylic acid
IX 2,3-dihydro-4-benzofuranmethanol
X 2,3-dihydro-4-benzofurancarboxaldehyde
XI Propanedioic acid
XII (E) -3 – (2,3-dihydro-4-benzofuranyl) propenoic acid
XIII thionyl chloride
XIV (E) -3 – (2,3-dihydro-4-benzofuranyl) propenoyl chloride
XV (3aS, 6R, 7aR)-hexahydro-8 ,8-dimethyl-3H-3a ,6-methano-2 ,1-benzisothiazole-2 ,2-dioxide
XVI (3aS,6R,7aR)-1-[(E)-3-(2,3-dihydro-4-benzofuranyl)-1-oxo-2-propenyl]hexahydro-8,8-dimethyl-3H-3a,6-methano-2,1-benzisothiazole-2,2-dioxide
XVII (3aS,6R,7aR)-1-[[(1R,2R)-2-(2,3-dihydro-4-benzofuranyl)cyclopropyl]carbonyl]hexahydro-8,8-dimethyl-3H-3a,6-methano-2,1-benzisothiazole-2,2-dioxide
XVIII [R-(R *, R *)] -2 – (2,3-dihydro-4-benzofuranyl) cyclopropanemethanol
XIX [R-(R *, R *)] -2 – (2,3-dihydro-4-benzofuranyl) cyclopropanecarboxaldehyde
XX hydroxylamine hydrochloride
XXI [R-(R *, R *)] -2 – (2,3-dihydro-4-benzofuranyl) cyclopropanecarbaldehyde oxime
XXII [R-(R *, R *)] -2 – (2,3-dihydro-4-benzofuranyl) cyclopropanemethanamine
XXIII propanoyl chloride
XXIV D-camphorsulfonic acid
XXV (1S, 4R) -7,7-dimethyl-2-oxo-bicyclo [2.2.1] heptane-1-methanesulfonyl chloride
XXVI ammonium hydroxide
XXVII (1S, 4R) -7,7-dimethyl-2-oxo-bicyclo [2.2.1] heptane-1-methanesulfonamide
XXVIII (3aS, 6R) -4,5,6,7-tetrahydro-8 ,8-dimethyl-3H-3a ,6-methano-2 ,1-benzisothiazole-2 ,2-dioxide
Bibliography
– Patents: Benzofuran and dihydrobenzofuran melatonergic agents: US5856529 (1999)
Priority: US19960032689P, 10 Dec. 1996 (Bristol-Myers Squibb Company, U.S.)
– Preparation III (quinazolines): US2004044015 (2004) Priority: EP20000402845, 13 Oct. 2000
– Preparation of VII (aminoalkylindols): Structure-Activity Relationships of Novel Cannabinoid Mimetics Eissenstat et al, J.. Med. Chem. 1995, 38, 3094-3105
– Preparation XXVIII: Towson et al. Organic Syntheses, Coll. Vol. 8, p.104 (1993) Vol. 69, p.158 (1990)
– Preparation XV: Weismiller et al. Organic Syntheses, Coll. Vol. 8, p.110 (1993) Vol. 69, p.154 (1990).
– G. Birznieks et al. Melatonin agonist VEC-162 Improves sleep onset and maintenance in a model of transient insomnia. Sleep 2007, 30, 0773 Abstract.
-. Rajaratnam SM et al, The melatonin agonist VEC-162 Phase time immediately advances the human circadian system, Sleep 2006, 29, 0159 Abstract.
-. AK Singh et al, Evolution of a manufacturing route for a highly potent drug candidate, 229th ACS Natl Meet, March 13-17, 2005, San Diego, Abstract MEDI 576.
– Vachharajani NN et al, Preclinical pharmacokinetics and metabolism of BMS-214778, a novel melatonin receptor agonist, J Pharm Sci. 2003 Apr; 92 (4) :760-72.
. – JW Scott et al, Catalytic Asymmetric Synthesis of a melotonin antagonist; synthesis and process optimization. 223rd ACS Natl Meet, April 7-11, Orlando, 2002, Abstract ORGN 186.
…………………….
SYNTHESIS CONSTRUCTION AS IN PATENT
GENERAL SCHEMES
Reaction Scheme 1

The syntheses of the 4-aryl-propenoic acid derivatives, 2 and 3, are shown in Reaction Scheme 1. The starting aldehydes, 1 , can be prepared by methods well known to those skilled in the art. Condensation of malonic acid with the aldehydes, 1, in solvents such as pyridine with catalysts such as piperidine or pyrrolidine, gives the 4-aryl- propenoic acid, 2. Subsequent conversion of the acid to the acid chloride using reagents such as thionyl chloride, phosphoryl chloride, or the like, followed by reaction with N,0-dimethyl hydroxylamine gives the amide intermediate 3 in good yields. Alternatively, aldehyde 1 can be converted directly to amide 3 using reagents such as diethyl (N-methoxy- N-methyl-carbamoylmethyl)phosphonate with a strong base such as sodium hydride.
Reaction Scheme 2

The conversion of the amide intermediate 3 to the racemic, trans- cyclopropane carboxaldehyde intermediate, 4, is shown in Reaction Scheme 2. Intermediate 3 was allowed to react with cyclopropanating reagents such as trimethylsulfoxonium iodide and sodium hydride in solvents such as DMF, THF, or the like. Subsequent reduction using reagents such as LAH in solvents such as THF, ethyl ether, or the like, gives the racemic, trans-cyclopropane carboxaldehyde intermediates, 4.
Reaction Scheme 3

Racemic cyclopropane intermediate 5 (R = halogen) can be prepared from intermediate 2 as shown in Reaction Scheme 3. Intermediate 2 was converted to the corresponding allylic alcohol by treatment with reducing agents such as sodium borohydride plus iodine in solvents such as THF. Subsequent acylation using reagents such as acetic anhydride in pyridine or acetyl chloride gave the allylic acetate which was allowed to react with cyclopropanating reagents such as sodium chloro-difluoroacetate in diglyme to provide the racemic, trans- cyclopropane acetate intermediates, 5. Reaction Scheme 4

The conversion of the acid 2 to the chiral cyclopropane carboxaldehyde intermediate, (-)-(trans)-4, is shown in Reaction Scheme 4. Intermediate 2 is condensed with (-)-2,10-camphorsultam under standard conditions, and then cyclopropanated in the presence of catalysts such as palladium acetate using diazomethane generated from reagents such as 1-methyl-3-nitro-1-nitrosoguanidine. Subsequent reduction using reagents such as LAH in solvents such as THF, followed by oxidation of the alcohol intermediates using reagents such as DMSO/oxalyl chloride, or PCC, gives the cyclopropane carboxaldehyde intermediate, (-)-(trans)-4, in good yields. The enantiomer, (+)-(trans)-4, can also be obtained employing a similar procedure using (+)-2,10- camphorsultam in place of (-)-2,10-camphorsultam.
When it is desired to prepare compounds of Formula I wherein m = 2, the alcohol intermediate may be activated in the conventional manner such as with mesyl chloride and treated with sodium cyanide followed by reduction of the nitrile group with a reducing agent such as LAH to produce the amine intermediate 6.
Reaction Scheme 5


Reaction Scheme 5 shows the conversion of intermediates 4 and 5 to the amine intermediate, 7, and the subsequent conversion of 6. or 7 to compounds of Formula I. The carboxaldehyde intermediate, 4, is condensed with hydroxylamine and then reduced with reagents such as LAH to give the amine intermediate, 7. The acetate intermediate 5 is hydrolyzed with potassium hydroxide to the alcohol, converted to the mesylate with methane sulfonyl chloride and triethyl amine in CH2CI2and then converted to the azide by treatment with sodium azide in solvents such as DMF. Subsequent reduction of the azide group with a reducing agent such as LAH produced the amine intermediate 7. Further reaction of 6 or 7 with acylating reagents gives compounds of Formula I. Suitable acylating agents include carboxylic acid halides, anhydrides, acyl imidazoles, alkyl isocyanates, alkyl isothiocyanates, and carboxylic acids in the presence of condensing agents, such as carbonyl imidazole, carbodiimides, and the like. Reaction Scheme 6

Reaction Scheme 6 shows the alkylation of secondary amides of Formula I (R2 = H) to give tertiary amides of Formula I (R2 = alkyl). The secondary amide is reacted with a base such as sodium hydride, potassium tert-butoxide, or the like, and then reacted with an alkylating reagent such as alkyl halides, alkyl sulfonate esters, or the like to produce tertiary amides of Formula I.
Reaction Scheme 7

Reaction Scheme 7 shows the halogenation of compounds of Formula I. The carboxamides, i (Q1 = Q2 = H), are reacted with excess amounts of halogenating agents such as iodine, N-bromosuccinimide, or the like to give the dihalo-compounds of Formula I (Q1 = Q2 = halogen). Alternatively, a stoichiometric amount of these halogenating agents can be used to give the monohalo-compounds of Formula I (Q1 = H, Q2 = halogen; or Q1 = halogen, Q2 = H). In both cases, additives such as lead IV tetraacetate can be used to facilitate the reaction. Biological Activity of the Compounds
The compounds of the invention are melatonergic agents. They have been found to bind human melatonergic receptors expressed in a stable cell line with good affinity. Further, the compounds are agonists as determined by their ability, like melatonin, to block the forskolin- stimulated accumulation of cAMP in certain cells. Due to these properties, the compounds and compositions of the invention should be useful as sedatives, chronobiotic agents, anxiolytics, antipsychotics, analgesics, and the like. Specifically, these agents should find use in the treatment of stress, sleep disorders, seasonal depression, appetite regulation, shifts in circadian cycles, melancholia, benign prostatic hyperplasia and related conditions
EXPERIMENTAL PROCEDURES
SEE ORIGINAL PATENT FOR CORECTIONS
Preparation 1
Benzofuran-4-carboxaldehyde
Step 1 : N-Methoxy-N-methyl-benzofuran-4-carboxamide
A mixture of benzofuran-4-carboxylic acid [Eissenstat, et al.. J. Medicinal Chemistry, 38 (16) 3094-3105 (1995)] (2.8 g, 17.4 mmol) and thionyl chloride (25 mL) was heated to reflux for 2 h and then concentrated in vacuo. The solid residue was dissolved in ethyl acetate (50 mL) and a solution of N,O-dimethylhydroxylamine hydrochloride (2.8 g) in saturated NaHC03(60 mL) was added with stirring. After stirring for 1.5 h, the ethyl acetate layer was separated. The aqueous layer was extracted with ethyl acetate. The ethyl acetate extracts were combined, washed with saturated NaHCO3 and concentrated in vacuo to give an oil (3.2 g, 95.4%).
Step 2: Benzofuran-4-carboxaldehyde
A solution of N-methoxy-N-methyl-benzofuran-4-carboxamide (3.2 g, 16.6 mmol) in THF (100 mL) was cooled to -45°C and then LAH (0.7 g, 18.7 mmol) was added. The mixture was stirred for 15 min, allowed to warm to -5°C, and then recooled to -45°C. Saturated KHS04 (25 mL) was added with vigorous stirring, and the mixture was allowed to warm to room temperature. The precipitate was filtered and washed with acetone. The filtrate was concentrated in vacuo to give an oil (2.3 g, 94%). Preparation 2
2,3-Dihydrobenzofuran-4-carboxaldehyde
Step 1 : 2,3-Dihydrobenzofuran-4-carboxylic acid
Benzofuran-4-carboxylic acid (10.0 g, 61 .7 mmol) was hydrogenated (60 psi) in acetic acid (100 mL) over 10% Pd/C (2 g) for 12 hr. The mixture was filtered and the filtrate was diluted with water (500 mL) to give 2,3- dihydrobenzofuran-4-carboxylic acid as a white powder (8.4 g, 83%). A sample was recrystallized from isopropanol to give fine white needles (mp: 185.5-187.5°C).
Step 2: (2,3-Dihydrobenzofuran-4-yl)methanol
A solution of 2,3-dihydrobenzofuran-4-carboxylic acid (10 g, 61 mmol) in THF (100 mL) was stirred as LAH (4.64 g, 122 mmol) was slowly added. The mixture was heated to reflux for 30 min. The mixture was cooled and quenched cautiously with ethyl acetate and then with 1 N HCI (150 mL). The mixture was then made acidic with 12 N HCI until all the inorganic precipitate dissolved. The organic layer was separated, and the inorganic layer was extracted twice with ethyl acetate. The organic layers were combined, washed twice with brine, and then concentrated in vacuo. This oil was Kϋgelrohr distilled to a clear oil that crystallized upon cooling (8.53 g, 87.6%).
Step 3: 2.3-Dihydrobenzofuran-4-carboxaldehyde
DMSO (8.10 mL, 1 14 mmol) was added at -78°C to a stirred solution of oxalyl chloride in CH2CI2 (40 mL of a 2M solution). A solution of (2,3- dihydrobenzofuran-4-yl)methanol (8.53 g, 56.9 mmol) in CH2CI2 (35 mL) was added dropwise, and the solution stirred at -78°C for 30 min. Triethyl amine (33 mL, 228 mmol) was added cautiously to quench the reaction. The resulting suspension was stirred at room temperature for 30 min and diluted with CH2CI2 (100 mL). The organic layer was washed three times with water, and twice with brine, and then concentrated in vacuo to an oil (8.42 g, 100%) that was used without purification.
Preparation 16
(±)-(trans)-2-(2,3-Dihyd robenzofuran-4-yl)cyclopropane- carboxaldehyde
Step 1 : (±Htrans)-N-Methoxy-N-methyl-2-(2.3-dihydrobenzofuran-4- yhcyclopropanecarboxamide
Trimethylsulfoxonium iodide (9.9 g, 45 mmol) was added in small portions to a suspension of sodium hydride (1 .8 g, 45 mmol) in DMF (120 mL). After the foaming had subsided (10 min), a solution of (trans)- N-methoxy-N-methyl-3-(2,3-dihydrobenzofuran-4-yl)propenamide (3.5 g, 15 mmol) in DMF (60 mL) was added dropwise, with the temperature maintained between 35-40°C. The mixture was stirred for 3 h at room temperature. Saturated NH4CI (50 mL) was added dropwise and the mixture was extracted three times with ethyl acetate. The organic extracts were combined, washed with H2O and brine, dried over K2CO3, and concentrated in vacuo to give a white wax (3.7 g, 100%).
Step 2: (±)-(trans)- 2-(2.3-Dihydrobenzofuran-4-yl)cyclopropane- carboxaldehyde
A solution of (±)-(trans)-N-methoxy-N-methyl-2-(2,3-dihydrobenzofuran- 4-yl)cyclopropanecarboxamide (3.7 g, 15 mmol) in THF (10 mL) was added dropwise to a rapidly stirred suspension of LAH (683 mg, 18 mmol) in THF (50 mL) at -45°C, maintaining the temperature below -40°C throughout. The cooling bath was removed, the reaction was allowed to warm to 5°C, and then the reaction was immediately recooled to -45°C. Potassium hydrogen sulfate (3.4 g, 25.5 mmol) in H20 (50 mL) was cautiously added dropwise, the temperature maintained below – 30°C throughout. The cooling bath was removed and the suspension was stirred at room temperature for 30 min. The mixture was filtered through Celite and the filter cake was washed with ether. The combined filtrates were then washed with cold 1 N HCI, 1 N NaOH, and brine. The filtrates were dried over MgSO4, and concentrated in vacuo to give a clear oil (2.6 g, 99%).
Preparation 18
(-)-(trans)-2-(2.3-Dihydrobenzofuran-4-yl)cyclopropane-carboxaldehyde
Step 1 : (-Htrans)-N-[3-(2.3-Dihvdrobenzofuran-4-yl)-propenoyll-2.10- camphorsultam
To a solution of (-)-2,10-camphorsultam (8.15 g, 37.9 mmol) in 50 mL toluene at 0°C was added sodium hydride (1.67 g, 41.7 mmol). After stirring for 0.33 h at 0°C and 0.5 h at 20°C and recooling to 0°C, a solution of 3-(2,3-dihydrobenzofuran-4-yl)-2-propenoyl chloride
(37.9 mmol), prepared in situ from the corresponding acid and thionyl chloride (75 mL), in toluene (50 mL), was added dropwise. After stirring for 18 h at 20°C, the mixture was diluted with ethyl acetate and washed with water, 1 N HCI, and 1 N NaOH. The organic solution was dried and concentrated in vacuo to give 15.8 g of crude product. Recrystallization form ethanol-methanol (600 mL, 1 :1) gave the product (13.5 g, 92%, mp 199.5-200°C).
Step 2: (-)-N-[[(trans)-2-(2,3-Dihydrobenzofuran-4-yl)-cyclopropylj- carbonylj-2, 10-camphorsultam
1 -Methyl-3-nitro-1 -nitrosoguanidine (23.88g 163 mmol) was added in portions to a mixture of 10 N sodium hydroxide (60 mL) and ether (200 mL) at 0°C. The mixture was shaken vigorously for 0.25 h and the ether layer carefully decanted into a solution of (-)-N-[3-(2,3-dihydrobenzofuran-4-yl)-2-propenoyl]-2,10-camphorsultam (9.67 g, 25 mmol) and palladium acetate (35 mg) in methylene chloride (200 mL). After stirring for 18 h, acetic acid (5 mL) was added to the reaction and the mixture stirred for 0.5 h. The mixture was washed with 1 N HCI, 1 N NaOH and brine. The solution was dried, concentrated in vacuo and the residue crystallized twice from ethanol to give the product (6.67 g, 66.5%, mp 157-159°C).
Step 3: (-)-(trans)-2-(2,3-Dihydrobenzofuran-4-yl)cyclopropane- methanol
A solution of (-)-N-[(trans)-2-(2,3-dihydrobenzofuran-4-yl)cyclo-propanecarbonylj-2,10-camphorsultam (4.3 g, 10.7 mmol) in THF (50 mL) was added dropwise to a mixture of LAH (0.81 g, 21.4 mmol) in THF (50 mL) at -45°C. The mixture was stirred for 2 hr while it warmed to 10°C. The mixture was recooled to -40°C and hydrolyzed by the addition of saturated KHS0 (20 mL). The mixture was stirred at room temperature for 30 minutes and filtered. The precipitate was washed twice with acetone. The combined filtrate and acetone washes were concentrated in vacuo. The gummy residue was dissolved in ether, washed with 1 N NaOH and 1 N HCI, and then dried in vacuo to give the product (2.0 g, 98.4%).
Step 4: (-)-(trans)-2-(2.3-Dihydrobenzofuran-4-yl)cyclopropane- carboxaldehyde DMSO (1.6 g, 21 mmol) was added to oxalyl chloride in CH2CI2(7.4 mL of 2 M solution, 14.8 mmole) at -78°C. The (-)-(trans)-2-(2,3-dihydrobenzofuran-4-yl)-cyclopropylmethanol (2.0 g, 10.5 mmol) in CH2CI2(15 mL) was added. The mixture was stirred for 20 min and then triethylamine (4.24 g, 42 mmol) was added. The mixture was warmed to room temperature and stirred for 30 min. The mixture was diluted with CH2CI2 and washed with water, 1 N HCI, and then 1 N NaOH. The organic layer was dried and concentrated iι> vacuo to give the aldehyde product (1.98 g, 100%).
Preparation 24
(-)-(trans)-2-(2.3-Dihydrobenzofuran-4-yl)cyclopropane-methanamine A mixture of (-)-(trans)-2-(2,3-dihydrobenzofuran-4-yl)cyclopropane-carboxaldehyde (1.98 g, 10.5 mmol), hydroxylamine hydrochloride (2.29 g, 33 mmol), and 30% NaOH (3.5 mL, 35 mmol), in 5:1
ethanol/water (50 mL) was heated on a steam bath for 2 h. The solution was concentrated in vacuo. and the residue mixed with water. The mixture was extracted with CH2CI2. The organic extracts were dried and concentrated in vacuo to give a solid which NMR analysis showed to be a mixture of the cis and trans oximes. This material was dissolved in THF (20 mL) and added to solution of alane in THF [prepared from LAH (1.14 g, 30 mmol) and H2S04 (1.47 g, 15 mmol) at 0°Cj. The reaction was stirred for 18 h, and quenched successively with water (1.15 mL), 15% NaOH (1.15 mL), and then water (3.45 mL). The mixture was filtered and the filtrate was concentrated in vacuo. The residue was mixed with ether and washed with water and then 1 N HCI. The acid washes were made basic and extracted with CH2CI . The extracts were dried and concentrated in vacuo to give the amine product (1.4 g, 70.5%). The amine was converted to the fumarate salt in ethanol (mp: 197-198°C).
Anal. Calc’d for C12H15NO • C4H404: C, 62.94; H, 6.27; N, 4.59.
Found: C, 62.87; H, 6.31 ; N, 4.52.
FINAL PRODUCT TASIMELTEON
Example 2
(-)-(trans)-N-[[2-(2,3-Dihydrobenzofuran-4-yl)cycloprop-1-yl]methyl]propanamide
This compound was prepared similar to the above procedure using propionyl chloride and (-)-(trans)-2-(2,3-dihydrobenzofuran-4-yl)- cyclopropanemethanamine to give an oil that solidified upon standing to an off-white solid (61 %, mp: 71-72°C). IR (NaCI Film): 3298, 1645, 1548, 1459, 1235 cm“1.
Mo5 : -17.3°
Anal. Calc’d for C15H19N02: C, 73.44; H, 7.87; N, 5.71 . Found: C, 73.28; H, 7.68; N, 5.58.
References
- ‘Time-bending drug’ for jet lag. BBC News. 2 December 2008
- Vachharajani, Nimish N., Yeleswaram, Krishnaswamy, Boulton, David W. (April 2003). “Preclinical pharmacokinetics and metabolism of BMS-214778, a novel melatonin receptor agonist”. Journal of Pharmaceutical Sciences 92 (4): 760–72. doi:10.1002/jps.10348. PMID 12661062.
- Shantha MW Rajaratnam, Mihael H Polymeropoulos, Dennis M Fisher, Thomas Roth, Christin Scott, Gunther Birznieks, Elizabeth B Klerman (2009-02-07). “Melatonin agonist tasimelteon (VEC-162) for transient insomnia after sleep-time shift: two randomised controlled multicentre trials”. The Lancet 373 (9662): 482–491. doi:10.1016/S0140-6736(08)61812-7. PMID 19054552. Retrieved 2010-02-23.
- Audio interview with Joseph Hull of Harvard, spring 2011
- Vanda Pharmaceuticals seeks FDA approval
- Recent progress in the development of agonists and antagonists for melatonin receptors.Zlotos DP.
Curr Med Chem. 2012;19(21):3532-49. Review.
7 Preclinical pharmacokinetics and metabolism of BMS-214778, a novel melatonin receptor agonist.
Vachharajani NN, Yeleswaram K, Boulton DW.J Pharm Sci. 2003 Apr;92(4):760-72.
 TASIMELTION
PATENTS
|
10-15-2010
|
PREDICTION OF SLEEP PARAMETER AND RESPONSE TO SLEEP-INDUCING COMPOUND BASED ON PER3 VNTR GENOTYPE
|
|
|
8-21-2009
|
TREATMENT FOR DEPRESSIVE DISORDERS
|
|
|
5-10-2000
|
Benzopyran derivatives as melatonergic agents
|
|
|
11-10-1999
|
Benzodioxa alkylene ethers as melatonergic agents
|
|
|
6-19-1998
|
BENZODIOXOLE, BENZOFURAN, DIHYDROBENZOFURAN, AND BENZODIOXANE MELATONERGIC AGENTS
|
| WO2007137244A1 * | May 22, 2007 | Nov 29, 2007 | Gunther Birznieks | Melatonin agonist treatment |
| US4880826 | Jun 25, 1987 | Nov 14, 1989 | Nava Zisapel | Melatonin antagonist |
| US4997845 | May 10, 1990 | Mar 5, 1991 | Eli Lilly And Company | β-alkylmelatonins as ovulation inhibitors |
| US5093352 | May 16, 1990 | Mar 3, 1992 | Whitby Research, Inc. | Antidepressant agents |
| US5151446 | Mar 28, 1991 | Sep 29, 1992 | Northwestern University | Substituted 2-amidotetralins as melatonin agonists and antagonists |
| US5225442 | Jan 3, 1992 | Jul 6, 1993 | Adir Et Compagnie | Compounds having a naphthalene structure |
| US5580878 | Jun 7, 1995 | Dec 3, 1996 | Interneuron Pharmaceuticals, Inc. | Substituted tryptamines phenalkylamines and related compounds |
| US5856529 | Dec 9, 1997 | Jan 5, 1999 | Bristol-Myers Squibb Company | Benzofuran and dihydrobenzofuran melatonergic agents |
| US6211225 | Jun 6, 2000 | Apr 3, 2001 | Bristol-Meyers Squibb | Heterocyclic aminopyrrolidine derivatives as melatonergic agents |
| US7754902 | May 18, 2006 | Jul 13, 2010 | Vanda Pharmaceuticals, Inc. | Ruthenium(II) catalysts for use in stereoselective cyclopropanations |
| US20010047016 | Apr 12, 2001 | Nov 29, 2001 | Gregory Oxenkrug | Method for treating depression |
| US20050164987 | Dec 22, 2004 | Jul 28, 2005 | Barberich Timothy J. | Melatonin combination therapy for improving sleep quality |
| US20090105333 | May 22, 2007 | Apr 23, 2009 | Gunther Birznieks | Melatonin agonist treatment |
extra info
|
- Department of Chemistry, Drexel University, Philadelphia, PA 19104.
- Shriner, R. L.; Shotton, J. A.; Sutherland, H. J. Am. Chem. Soc. 1938, 60, 2794.
- Oppolzer, W.; Chapuis, C.; Bernardinelli, G. Helv. Chim. Acta 1984, 67, 1397.
- Vandewalle, M.; Van der Eycken, J.; Oppolzer, W.; Vullioud, C. Tetrahedron 1986, 42, 4035.
- Davis, F. A.; Towson, J. C.; Weismiller, M. C.; Lal, G.; Carroll,, P. J. J. Am. Chem. Soc. 1988, 110, 8477.
- Oppolzer, W. Tetrahedron 1987, 43, 1969.
- Oppolzer, W.; Mills, R. J.; Pachinger, W.; Stevenson, T. Helv. Chim. Acta 1986, 69, 1542; Oppolzer, W.; Schneider, P. Helv. Chim. Acta 1986, 69, 1817; Oppolzer, W.; Mills, R. J.; Réglier, M. Tetrahedron Lett. 1986, 27, 183; Oppolzer, W.; Poli. G.Tetrahedron Lett. 1986, 27, 4717; Oppolzer, W.; Poli, G.; Starkemann, C.; Bernardinelli, G. Tetrahedron Lett. 1988, 29, 3559.
- Oppolzer, W.; Barras, J-P. Helv. Chim. Acta 1987, 70, 1666.
- Curran, D. P.; Kim, B. H.; Daugherty, J.; Heffner, T. A. Tetrahedron Lett. 1988, 29, 3555.
- Differding, E.; Lang, R. W. Tetrahedron Lett. 1988, 29, 6087.
Org. Synth. 1990, 69, 158 |
- Org. Syn. Coll. Vol. 8, 110
- Org. Syn. Coll. Vol. 9, 212
-
References and Notes
- Department of Chemistry, Drexel University, Philadelphia, PA 19104.
- Reychler, M. A. Bull. Soc. Chim. III 1889, 19, 120.
- Armstrong, H. E.; Lowry, T. M. J. Chem. Soc., Trans. 1902, 81, 1441.
- Dauphin, G.; Kergomard, A.; Scarset, A. Bull. Soc. Chim. Fr. 1976, 862.
- Davis, F. A.; Jenkins, Jr., R. H.; Awad, S. B.; Stringer, O. D.; Watson, W. H.; Galloy, J. J. Am. Chem. Soc. 1982, 104, 5412.
- Vandewalle, M.; Van der Eycken, J.; Oppolzer, W.; Vullioud, C. Tetrahedron, 1986, 42, 4035.
- Davis, F. A.; Towson, J. C.; Weismiller, M. C.; Lal, S.; Carroll, P. J. J. Am. Chem. Soc. 1988, 110, 8477.
- Davis, F. A.; Weismiller, M. C.; Lal, G. S.; Chen, B. C.; Przeslawski, R. M. Tetrahedron Lett., 1989, 30, 1613.
- Oppolzer, W. Tetrahedron 1987, 43, 1969.
- Glahsl, G.; Herrmann, R. J. Chem. Soc., Perkin Trans. I 1988, 1753.
- Differding, E.; Lang, R. W. Tetrahedron Lett. 1988, 29, 6087.
- For recent reviews on the chemistry of N-sulfonyloxaziridines, see: (a) Davis, F. A.; Jenkins, Jr., R. H. in “Asymmetric Synthesis,” Morrison, J. D., Ed.; Academic Press: Orlando, FL, 1984, Vol. 4, Chapter 4;
- Davis, F. A.; Haque, S. M. in “Advances in Oxygenated Processes,” Baumstark, A. L., Ed.; JAI Press: London, Vol. 2;
- Davis, F. A.; Sheppard, A. C. Tetrahedron 1989, 45, 5703.
- Davis, F. A.; McCauley, Jr., J. P.; Chattopadhyay, S.; Harakal, M. E.; Towson, J. C.; Watson, W. H.; Tavanaiepour, I. J. Am. Chem. Soc. 1987, 109, 3370.
- Davis, F. A.; Stringer, O. D.; McCauley, Jr., J. M. Tetrahedron 1985, 41, 4747.
- Davis, F. A.; Chattopadhyay, S. Tetrahedron Lett. 1986, 27, 5079.
- Davis, F. A.; Harakal, M. E.; Awad, S. B. J. Am. Chem. Soc. 1983, 105, 3123.
- Davis, F. A.; Wei, J.; Sheppard, A. C.; Gubernick S. Tetrahedron Lett. 1987, 28, 5115.
- Davis, F. A.; Lal, G. S.; Wei, J. Tetrahedron Lett. 1988, 29, 4269.
- Davis, F. A.; Haque, M. S.; Ulatowski, T. G.; Towson, J. C. J. Org. Chem. 1986, 51, 2402.
- Davis, F. A.; Haque, M. S. J. Org. Chem. 1986, 51, 4083; Davis, F. A.; Haque, M. S.; Przeslawski, R. M. J. Org. Chem. 1989, 54, 2021.
- Davis, F. A.; Ulatowski, T. G.; Haque, M. S. J. Org. Chem. 1987, 52, 5288.
- Davis, F. A.; Sheppard, A. C., Lal, G. S. Tetrahedron Lett. 1989, 30, 779.
- Davis, F. A.; Sheppard, A. C.; Chen, B. C.; Haque, M. S. J. Am. Chem. Soc. 1990, 112, 6679.

dedicated to lionel my son


my daughter Aishal

THEY KEEP ME GOING
India’s GVK BIO goes international with Aragen deal

GVK Biosciences (GVK BIO), the discovery research and development organisation based in Hyderabad, India, has reached a definitive agreement to acquire Aragen Bioscience, a preclinical contract research organisation operating out of Morgan Hill, US and specialising in high-value biologics services.
No financial terms were disclosed for the deal, in which GVK BIO is acquiring the capital stock of Aragen Bioscience. It is the Indian company’s first international acquisition.
According to a report in India’s Business Standard, GVK is taking a 65% stake in Aragen Biosciences and will acquire the remaining 35% over a period of two years, leaving Aragen to function as a separate entity.
Read more at: http://www.pharmatimes.com/Article/14-01-30/India_s_GVK_BIO_goes_international_with_Aragen_deal.aspx#ixzz2s3enmHuZ
Follow us: @PharmaTimes on Twitter
Lodenafil Carbonate … an Erectile Dysfunction Drug in Phase III

Lodenafil carbonate
UNII-29X84F932D, CRIS-031
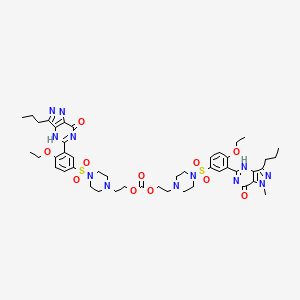
bis-(2-{4-[4-ethoxy-3-(1-methyl-7-oxo-3-propyl-6,7-dihydro-1H-pyrazolo[4,3-d]pyrimidin-5-yl)-benzenesulfonyl]piperazin-1-yl}-ethyl)carbonate
5-{2-ethoxy-5-[(4-hydroxyethyl-1-piperazinyl)sulfonyl]phenyl}-1-methyl-3-n-propyl-1,6-dihydro-7H-pyrazole[4,3-d]pyrimidin-7-one. IS THE NAME OF MONOMER
398507-55-6 CAS
Cristalia (Originator)
| C47 H62 N12 O11 S2= MF | |
| Molecular Weight | 1035.199 |
Lodenafil is a drug belonging to a class of drugs called PDE5 inhibitor, which many other erectile dysfunction drugs such as sildenafil, tadalafil, and vardenafil also belong to. Like udenafil and avanafil it belongs to a new generation of PDE5 inhibitors.
- Sildenafil, tadalafil, vardenafil, and the newer udenafil and avanafil selectively inhibit PDE5, which is cGMP-specific and responsible for the degradation of cGMP in the corpus cavernosum. These phosphodiesterase inhibitors are used primarily as remedies for erectile dysfunction, as well as having some other medical applications such as treatment of pulmonary hypertension.
Lodenafil is formulated as a dimer, lodenafil carbonate, which breaks down in the body to form two molecules of the active drug lodenafil. This formulation has higher oral bioavailability than the parent drug.[1]
It is manufactured by Cristália Produtos Químicos e Farmacêuticos in Brazil and sold there under the brand-name Helleva.[2]

Helleva (Lodenafil Carbonate) is an oral PDE5 inhibitor prescribed to treat men suffering from erectile dysfunction. It operates by relaxing muscles and dilating blood vessels in the penis to increase circulation making it easier to attain and maintain an erection.
It has undergone Phase III clinical trials,[3][4][5] but is not yet approved for use in the United States by the U.S. Food and Drug Administration.
………..
SYNTHESIS
WO 2002012241 OR US7148350
MONOMER synthesis
 PIPERAZINE
AND
 ETHYL CHLORO ACETATE
WILL GIVE
Ethyl 1-piperazinylacetate
SEE RXN 1 BELOW
Reaction 1:
Synthesis of Piperazine Ethyl Acetate
To a reaction blend containing 100 g (3 Eq, 0.515 mol, MW=194) of piperazine, 26.3 mL (1.1 Eq, 0.189 mol, MW=101, d=0.726) of triethylamine in 200 mL of isopropanol, add to a solution previously prepared of 18.4 mL (1 Eq., 0.172 mol, MW=122.55, d=1.15) of chloroacetate of ethyl in 140 mL of isopropanol under stirring, at room temperature. Keep the reaction medium under stirring, monitoring the reaction termination by means of a chromatography of the thin layer (about 2–3 hours). Add a solution of 40.6 g (0.344 mol) of succinic acid in 140 mL of isopropanol. Keep the system under stirring for about 30 minutes to assure total precipitation of the succinate salt of piperazine formed. Filter this salt and concentrate the filtrate containing the mono and dialkyled derivatives. We obtain a slightly yellowish oil, which is used in later phases without purification.
Mass obtained=33 g
GC/MS: Monoalkylated derivative 72%, and dialkylated 22%.
NEXT
Piperazine Ethyl Acetate
AND

5-(5-chlorosulfonyl-2-ethoxyphenyl)-1-methyl-3n-propyl-1,6-dihydro-7H-pyrazole[4,3-d]pyrimidin-7-one
WILL REACT TO GIVE… 5-{2-ethoxy-5-[(4-ethyl acetate 1-piperazinyl)sulfonyl]phenyl}-1-methyl-3-n-propyl-1,6-di-hydro-7H-pyrazole[4,3-d]pyrimidin-7-one AS IN RXN 4 BELOW
Reaction 4:
Synthesis of 5-{2-ethoxy-5-[(4-ethyl acetate 1-piperazinyl)sulfonyl]phenyl}-1-methyl-3-n-propyl-1,6-di-hydro-7H-pyrazole[4,3-d]pyrimidin-7-one.
Suspend 24.6 g (60 mmol, MW=410.9) of 5-(5-chlorosulfonyl-2-etoxyphenyl)-1-methyl-3n-propyl-1,6-dihydro-7H-pyrazole[4,3-d]pyrimidin-7-one in 900 mL of ethanol absolute. Under stirring and at room temperature, add at only one time, a solution containing 31.0 g (3 Eq., 180 mmol MW=172) of N-piperazine ethyl acetate (Reaction 1) dissolved in 150 mL of ethanol absolute. In an interval of 2–10 minutes, all solid is consumed, forming a clean and homogeneous solution, and after that starts the precipitation of the expected product. At the end of the reaction, which lasts 2–3 hours (monitored by chromatography of thin layer), the product is vacuum filtered and the solid is washed with two portions of 50 mL of iced absolute ethanol. 29 g are obtained (yielding=89%) from the product as a white solid of MP=165.5–166.5° C.
Reaction 7:
Intermediate 1
5-{2-ethoxy-5-[(4-hydroxyethyl-1-piperazinyl)sulfonyl]phenyl}-1-methyl-3-n-propyl-1,6-dihydro-7H-pyrazole[4,3-d]pyrimidin-7-one. IS MONOMER
please note during LAH redn …………. the PIP CH2-C=O-O CH2 CH3 BECOMES PIP-CH2CH2-OH
To a suspension of lithium aluminum hydride (0.74 g 2.2 Eq. MW=37.9) in 25 mL of THF, slowly add, under stirring and at room temperature, a suspension of 5.0 g (9.1 mmol, MW=546.6) of 5-{2-ethoxy-5-[(4-ethyl acetate 1-piperazinyl)sulfonyl]phenyl}-1-methyl-3-n-propyl-1,6-di-hydro-7H-pyrazole[4,3-d]pyrimidin-7-one in 50 mL of THF. The system is maintained under stirring, monitoring the consumption of the product by chromatography of thin layer, until the complete consumption of the starting reagent (about 5–6 hours). Slowly add water to the reaction medium and, when there is no longer release of H2, add HCl 1M regulating pH for 7. Extract the product with 3 200 mL-portions of chloroform, dry with anhydrous sodium sulfate and vacuum concentrate the product. It is obtained 3.8 g of the product as a cream solid MP=183–187° C. yielding 83%. The same was crystallized from methanol and DMF yielding a slightly yellowish solid with melting point at 189–192° C.

note …………. the PIP CH2-C=O-O CH2 CH3 BECOMES PIP-CH2CH2-OH
HOMODIMER CARBONATE
EXAMPLE 1B
Homodimer Carbonate of Intermediate 1—Alternative Method
A phosgene solution (3.5 g, 35 mmol) dissolved in 20 mL of toluene was added dropwise to a solution of 2.02 g (4 mmol) of 5-{2-ethoxy-5-[(4-hydroxyethyl-1-piperazinyl)sulfonyl]phenyl}-1-methyl-3-n-propyl-1,6-dihydro-7H-pyrazole[4,3-d]pyrimidin-7-one, suspended in 44 mL of toluene. The reaction mixture resulting is stirred and followed by chromatography analysis of thin layer every hour until the reagent conversion in its chloroformate was completed. When the analysis indicates the complete consumption of 5-{2-ethoxy-5-[(4-hydroxyethyl-1-piperazinyl)sulfonyl]phenyl}-1-methyl-3-n-propyl-1,6-dihydro-7H-pyrazole[4,3-d]pyrimidin-7-one, the volatile compounds of the reaction are vacuum removed (solvents and phosgene), yielding the esther chloroformate raw derivative of 5-{2-ethoxy-5-[(4-hydroxyethyl-1-piperazinyl)sulfonyl]phenyl}-1-methyl-3-n-propyl-1,6-dihydro-7H-pyrazole[4,3-d]pyrimidin-7-one.
The raw chloroformate obtained above (4.0 mmol, 2.27 g) is dissolved in about 30 mL of dichloromethane, to which is added 2.07 g (4.1 mmol) of 5-{2-ethoxy-5-[(4-hydroxyethyl-1-piperazinyl)sulfonyl]phenyl}-1-methyl-3-n-propyl-1,6-dihydro-7H-pyrazole[4,3-d]pyrimidin-7-one, followed by the addition of 4 mL of dichloromethane containing 450 mg of triethylamine. The reaction mixture is maintained under stirring, being followed by chromatography of thin layer every hour until this indicates the end of the reaction (disappearing of chloroformate derivative). The reaction mixture is then diluted with 60 mL of dichloromethane, washed with NaCl saturated solution, after with sodium bicarbonate saturated solution and again with NaCl saturated solution. Organic phase is separated and dry with anhydrous sodium sulfate. The solvent is then evaporated to dry, yielding the dimer carbonate as a slightly yellowish solid.
This compound is re-crystallized from ethanol:DMF, yielding a pale white solid. Yielding m=3.2 g (76%)
Microanalysis: Theoretical C, (54.53%); H, (6.04%); N, (16.24%);
Obtained C, (54.45%); H, (6.02%); N, (16.17%).
INFO ABOUT INTERMEDIATE
5-(5-chlorosulfonyl-2-ethoxyphenyl)-1-methyl-3n-propyl-1,6-dihydro-7H-pyrazole[4,3-d]pyrimidin-7-one
| CAS No. | 139756-22-2 |
| Chemical Name: | 5-(5-Chlorosulfonyl-2-ethoxyphenyl)-1-methyl-3-propyl-1,6-dihydro-7H-pyrazolo[4,3-d]pyrimidin-7-one |
| Synonyms: | Sildenafil Chlorosulfone IMpurity;Sildenafil Chlorosulfonyl IMpurity;5-(5-CHLOROSULFONYL-2-ETHOXY PHENYL)-1-METHYL-3-N-PROPYL-1;3-(6,7-dihydro-1-methyl-7-oxo-3-propyl-1 H-pyrazolo-(4-3-d)-pyrimidine-5;5-(5-Chlorosulfonyl-2-ethoxyphenyl)-1-methyl-3-propyl-1,6-dihydro-7H-pyrazolo[4,3-d]pyrimidin-7-one;3-(4,7-Dihydro-1-methyl-7-oxo-3-propyl-1H-pyrazolo[4,3-d]pyrimidin-5-yl)-4-ethoxy-benzenesulfonyl Chloride;4-Ethoxy-3-(1-Methyl-7-oxo-3-propyl-6,7-dihydro-1H-pyrazolo[4,3-d]pyriMidin-5-yl)benzene-1-sulfonyl chloride |
| CBNumber: | CB11175931 |
| Molecular Formula: | C17H19ClN4O4S |
http://www.chemicalbook.com/ChemicalProductProperty_EN_CB11175931.htm
…………..
SYNTHESIS OF

http://www.google.co.in/patents/US6362178

22.27 g (250 mmol) of D,L-alanine and 55.66 g (550 mmol) of triethylamine are dissolved in 250 ml of dichloromethane, and the solution is cooled to 0° C. 59.75 g (550 mmol) of trimethylsilyl chloride are added dropwise, and the solution is stirred for 1 hour at room temperature and for 1 hour at 40° C. After cooling to −10° C., 26.64 g (250 mmol) of butyryl chloride are added dropwise, and the resulting mixture is stirred for 2 hours at −10° C. and for one hour at room temperature.
With ice-cooling, 125 ml of water are added dropwise and the reaction mixture is stirred at room temperature for 15 minutes. The aqueous phase is evaporated to dryness, the residue is titrated with acetone and the mother liquor is filtered off with suction. The solvent is removed and the residue is chromatographed. The resulting product is dissolved in 3N aqueous sodium hydroxide solution and the resulting solution is evaporated to dryness. The residue is taken up in conc. HCl and once more evaporated to dryness. The residue is stirred with acetone, precipitated solid is filtered off with suction and the solvent is removed under reduced pressure. This gives 28.2 g (71%) of a viscous oil which crystallizes after some time.
200 MHz 1H-NMR (DMSO-d6): 0.84, t, 3H; 1.22, d, 3H; 1.50, hex, 2H; 2.07, t, 2H; 4.20, quin., 1H; 8.09, d, 1H.

25 g (210 mmol) of 2-hydroxybenzonitrile are refluxed with 87 g of potassium carbonate and 34.3 g (314.8 mmol) of ethyl bromide in 500 ml of acetone overnight. The solid is filtered off, the solvent is removed under reduced pressure and the residue is distilled under reduced pressure. This gives 30.0 g (97%) of a colourless liquid.
200 MHz 1H-NMR (DMSO-d6): 1.48, t, 3H; 4.15, quart., 2H; 6.99, dt, 2H; 7.51, dt, 2H.

21.4 g (400 mmol) of ammonium chloride are suspended in 375 ml of toluene, and the suspension is cooled to 0° C. 200 ml of a 2M solution of trimethylaluminium in hexane are added dropwise, and the mixture is stirred at room temperature until the evolution of gas has ceased. After addition of 29.44 g (200 mmol) of 2-ethoxybenzonitrile, the reaction mixture is stirred at 80° C. (bath) overnight.
With ice-cooling, the cooled reaction mixture is added to a suspension of 100 g of silica gel and 950 ml of chloroform, and the mixture is stirred at room temperature for 30 minutes. The mixture is filtered off with suction, and the filter residue is washed with the same amount of methanol. The mother liquor is concentrated, the resulting residue is stirred with a mixture of dichloromethane and methanol (9:1), the solid is filtered off with suction and the mother liquor is concentrated. This gives 30.4 g (76%) of a colourless solid.
200 MHz 1H-NMR (DMSO-d6): 1.36, t, 3H; 4.12, quart., 2H; 7.10, t, 1H; 7.21, d, 1H; 7.52, m, 2H; 9.30, s, broad, 4H.
EXAMPLE 10A 2-(2-Ethoxy-phenyl)-5-methyl-7-propyl-3H-imidazo[5,1-f][1,2,4]triazin-4-one

7.16 g (45 mmol) of 2-butyrylamino-propionic acid and 10.67 g of pyridine are dissolved in 45 ml of THF and, after addition of a spatula tip of DMAP, heated to reflux. 12.29 g (90 mmol) of ethyl oxalyl chloride are slowly added dropwise, and the reaction mixture is refluxed for 3 hours. The mixture is poured into ice-water and extracted three times with ethyl acetate and the organic phase is dried over sodium sulphate and concentrated using a rotary evaporator. The residue is taken up in 15 ml of ethanol and refluxed with 2.15 g of sodium bicarbonate for 2.5 hours. The cooled solution is filtered.
With ice-cooling, 2.25 g (45 mmol) of hydrazine hydrate are added dropwise to a solution of 9.03 g (45 mmol) of 2-ethoxybenzamidine hydrochloride in 45 ml of ethanol, and the resulting suspension is stirred at room temperature for another 10 minutes. The ethanolic solution described above is added to this reaction mixture, and the mixture is stirred at a bath temperature of 70° C. for 4 hours. After filtration, the mixture is concentrated, the residue is partitioned between dichloromethane and water, the organic phase is dried over sodium sulphate and the solvent is removed under reduced pressure.
This residue is dissolved in 60 ml of 1,2-dichloroethane and, after addition of 7.5 ml of phosphorus oxychloride, refluxed for 2 hours. The mixture is diluted with dichloromethane and neutralized by addition of sodium bicarbonate solution and solid sodium bicarbonate. The organic phase is dried and the solvent is removed under reduced pressure. Chromatography using ethyl acetate and crystallization afford 4.00 g (28%) of a colourless solid, Rf=0.42 (dichloromethane/methanol=95:5)
200 MHz 1H-NMR (CDCl3): 1.02, t, 3H; 1.56, t, 3H; 1.89, hex, 2H; 2.67, s, 3H; 3.00, t, 2H; 4.26, quart., 2H; 7.05, m, 2H; 7.50, dt, 1H; 8.17, dd, 1H; 10.00, s, 1H.
EXAMPLE 15A 4-Ethoxy-3-(5-methyl-4-oxo-7-propyl-3,4-dihydro-imidazo[5,1-f][1,2,4]triazin-2-yl)-benzenesulphonyl chloride

At 0° C., 2.00 g (6.4 mmol) of 2-(2-ethoxy-phenyl)-5-methyl-7-propyl-3H-imidazo[5,1-f][1,2,4]triazin-4-one are slowly added to 3.83 ml of chlorosulphonic acid. At room temperature, the reaction mixture is stirred ovemight, and then poured into ice-water and extracted with dichloromethane. This gives 2.40 g (91%) of a colourless foam.
200 MHz 1H-NMR (CDCl3): 1.03, t, 3H; 1.61, t, 2H; 1.92, hex, 2H; 2.67, s, 3H; 3.10, t, 2H; 4.42, quart., 2H; 7.27, t, 1H; 8.20, dd, 1H; 8.67, d, 1H; 10.18, s, 1H.
Example 22 2-[2-Ethoxy-5-(4-hydroxyethyl-1-amino-piperazine-1-sulphonyl)-phenyl]-5-methyl-7-propyl-3H-imidazo[5,1-f][1,2,4]triazin-4-one
By the same method, starting with 0.04 g (0.097 mmol) of 4-ethoxy-3-(5-methyl-4-oxo-7-propyl-3,4-dihydro-imidazo[5,1-f][1,2,4]triazin-2-yl)-benzenesulphonyl chloride and 0.04 g (0.29 mmol) of 1-amino-4-hydroxyethylpiperazine, 46 mg (91%) of 2-[2-ethoxy-5-(4-hydroxyethyl-1-amino-piperazine-1-sulphonyl)-phenyl]-5-methyl-7-propyl-3H-imidazo[5,1-f][1,2,4]triazin-4-one are obtained.
Rf=0.08 (dichloromethane/methanol=19:1)
200 MHz 1H-NMR (CDCl3): 1.02, t, 3H; 1.59, t, 3H; 1.90, sex., 2H; 2.49, m, 6H; 2.62, s, 3H; 2.71, m, 4H; 3.00, t, 2H; 3.55, t, 2H; 4.31, quart., 2H; 7.14, d, 1H; 8.05, dd, 1H; 8.60, d, 1H.
…………..
Methods of analysis
The development of lodenafil carbonate was reported by Toque et al. (2008). They observed the effects of lodenafil carbonate on rabbit and human corpus cavernosum relaxation, activity of PDE5 in human platelets, stability and metabolic studies in comparison with sildenafil and lodenafil, as well as the pharmacological evaluation of lodenafil carbonate after intravenous and oral administration in male beagles.
The determination of PDE activity, stability of lodenafil carbonate in human, dog and rat plasma and the pharmacokinetic parameters after a single intravenous or oral dose was carried out by LC-MS/MS analysis
Codevilla et al. (2011a) developed a stability-indicating reversed-phase liquid chromatography method using ultraviolet (UV) detection for the quantitative determination of lodenafil carbonate in tablets. The method can be useful for routine quality control assay and stability studies.
Another study for the determination of lodenafil carbonate in tablets was developed by Codevilla et al. (2011b). As an alternative to the LC method the authors suggested a UV-spectrophotometric method for the analysis of lodenafil carbonate in pharmaceutical form. The UV method offers advantages over other analytical methods due to its rapidity, simplicity, and lower cost. Recently, Codevilla et al. (2012) developed and validated a capillary zone electrophoresis (CZE) method for determination of lodenafil carbonate in drug products. There are some advantages to use the CZE method, such as rapid analysis, small sample and reagent consumption, high separation efficiency (Furlanetto et al., 2001; Yang et al., 2010). The results obtained from the UV-spectrophotometric method and CZE method were compared statistically with the LC method (Codevilla et al., 2011a) and the results showed no significant difference between these methods.
References
- Toque HA, Teixeira CE, Lorenzetti R, Okuyama CE, Antunes E, De Nucci G (September 2008). “Pharmacological characterization of a novel phosphodiesterase type 5 (PDE5) inhibitor lodenafil carbonate on human and rabbit corpus cavernosum”. European Journal of Pharmacology 591 (1–3): 189–95. doi:10.1016/j.ejphar.2008.06.055. PMID 18593576.
- Cristália Product page. Retrieved on September 16, 2009.
- ukmedix Lodenafil article. Retrieved on September 16, 2009.
- Glina S, Toscano I, Gomatzky C, de Góes PM, Júnior AN, Claro JF, Pagani E (February 2009). “Efficacy and tolerability of lodenafil carbonate for oral therapy in erectile dysfunction: a phase II clinical trial”. The Journal of Sexual Medicine 6 (2): 553–7. doi:10.1111/j.1743-6109.2008.01079.x.PMID 19040623.
- Glina S, Fonseca GN, Bertero EB, Damião R, Rocha LC, Jardim CR, Cairoli CE, Teloken C, Torres LO, Faria GE, da Silva MB, Pagani E (February 2010). “Efficacy and Tolerability of Lodenafil Carbonate for Oral Therapy of Erectile Dysfunction: A Phase III Clinical Trial”. The Journal of Sexual Medicine 7 (5): 1928–1936. doi:10.1111/j.1743-6109.2010.01711.x. PMID 20214718.
- Toque H A et al., (2008) European Journal of Pharmacology, 591(1-3):189-95.
- Exploring the role of PDE5 inhibition in the treatment of muscular dystrophy
Drugs Fut 2011, 36(4): 321
VARDENAFIL

GSK’s diabetes drug Eperzan moves towards approval in Europe
GlaxoSmithKline has received a positive opinion from the European Medicines Agency’s (EMA) Committee for Medicinal Products for Human Use (CHMP) for albiglutide, under the brand name Eperzan, for treatment of type 2 diabetes.
CLICK ON TITLE
GSK’s diabetes drug Eperzan moves towards approval in Europe
Mirodenafil 米罗那非 标准品 ………..An erectogenic agent.

862189-95-5 (free base)
SK Chemicals (Originator)
Treatment of Erectile Dysfunction , hypertention
Mirodenafil belongs to a class of drugs called PDE5 inhibitors, which many other erectile dysfunction drugs such as sildenafil, tadalafil, andvardenafil also belong to. It was developed by SK Chemicals Life Science and is marketed under the trade name of Mvix tab which comes in different doses (50 mg, 100 mg).
Mirodenafil is also available under the name of Mvix S ODF 50 mg as an orally dissolving film (ODF) which dissolves on the tongue without water. It is the first licensed medicine for the treatment of erectile dysfunction as a dosage form of film.
Mirodenafil is a newly developed oral phosphodiesterase type 5 inhibitor, currently under investigation as a treatment for erectile dysfunction (ED).
MIRODENAFIL米罗那非 标准品
Mirodenafil hydrochloride is a high selective PDE5 inhibitor commercialized by SK Chemicals which had been in early clinical development for the treatment of erectile dysfunction (ED). Early clinical studies had also been ongoing for the treatment of hypertension in patients taking amlodipine; however, no recent development has been reported for this research. The development of compound started in 1998 jointly by SK Chemicals and a bio-venture In2Gen.
Several clinical trials were conducted,[1][2][3] but mirodenafil has not been approved for use in the United States by the U.S. Food and Drug Administration.

Mirodenafil dihydrochloride
|
||||||||||||
Korean Patent No. 358083 discloses pyrrolopyrimidinone derivatives having good inhibition activity against PDE-5, a method of its preparation thereof, an intermediate compound used to prepare the same and their use for prevention and treatment of erectile dysfunction, pulmonary arterial hypertension, chronic obstructive pulmonary disease, benign prostatic hypertrophy and lower urinary tract diseases.
Of the pyrrolopyrimidinone derivatives disclosed in Korean Patent No. 358083, 5-ethyl-2-{5-[4- (2-hydroxyethyl)piperazin-1-ylsulfonyl]-2-n-propoxyphenyl}-7-n-propyl-l-3,5-dihydro-4 H-pyrrolo[3,2-d]pyrimidin-4-one (hereinafter, “SK-3530”) represented by the following formula (1 ) is an excellent selective inhibitor PDE-5 over other PDEs and is under clinical trial for the treatment of erectile dysfunction after passing through the preclinical stage.

The dihydrochloride salt (2HCI) of SK-3530 has been under investigation through the preclinical and clinical stages.
The SK-3530 dihydrochloride salt has good solubility and can be easily stabilized for pharmaceutical preparation. But, it has the following drawbacks.
First, because the SK-3530 dihydrochloride salt is hygroscopic, it easily absorbs moisture from the atmosphere and becomes discolored when the moisture content is high. And, due to the hygroscopic property, an anhydrous solvent condition and a dry air condition have to be provided to obtain a stable product. Second, the SK-3530 dihydrochloride salt should be kept at a temperature lower than room temperature because it does not show enough stability at room temperature. In particular, the SK-3530 dihydrochloride salt is labile to heat or light, and thus any prolonged exposure to heat or light results in various impurities.
Third, the SK-3530 dihydrochloride salt could corrode the punch during tablet ting due to its somewhat corrosive properties. This is because the SK-3530 dihydrochloride salt is a simple amorphous salt rather than being a stable crystalline acid addition salt or hydrate form. Thus, one of the two hydrochloric acid groups with a relatively weak ionic bond character may leave the molecule under severe conditions. As aforementioned, the SK-3530 dihydrochloride salt may be endowed with a sufficient stability for pharmaceutical preparation. But, some additional techniques and costs are needed due to the deficiency in intrinsic physicochemical property and stability of the compound.


…………………………

The invention relates to a series of pyrrolopyrimidinone derivatives of the formula (1):

R1 ETHYL
R2=H
R3= PROPYL
R4 = PROPYL
R5=R5=SO2NR6R7, NR6R7 is 4-(3-hydroxypropyl)piperazinyl) IS MIRODENAFIL
ANALOGOUS METHOD
BELOW IS CUT PASTE OF R1 METHYL ANALOGUE ……………..R1 =METHYL AND NOT ETHYL ….CAUTION
Example 39 Preparation of
5-(5-(4-(2-hydroxyethyl)piperazinylsulfonyl)-2-n-propoxyphenyl)-1-methyl-3-n-propyl-1,6-dihydro-7H-pyrrolo[4,3-d]pyrimidin-7-one hydrochloride (a compound of the formula (1) wherein R5=SO2NR6R7, R1=CH3, R2=H, R3=CH2CH2CH3, R4=CH2CH2CH3; NR6R7 is 4-(2-hydroxyethyl)piperazinyl)
The titled compound was prepared as described in Example 2 by using 5-(5-(4-(2-hydroxyethyl)piperazinylsulfonyl)-2-n-propoxyphenyl)-1-methyl-3-n-propyl-1,6-dihydro-7H-pyrrolo[4,3-d]pyrimidin-7-one in place of 5-(2-ethoxy-5-(4-methylpiperazinylsulfonyl)phenyl)-1-methyl-3-n-propyl-1,6-dihydro-7H-pyrrolo[4,3-d]pyrimidin-7-one.
yield: 99%
mp 66.5° C. dec;
IR (neat) 3332 (NH and OH), 1676 (C═O), 1166 (SO2) cm−1;
1H NMR (DMSO-d6) δ 0.92 (t, J=7.2 Hz, 3H, CH2CH2CH3), 0.96 (t, J=7.2 Hz, 3H, OCH2CH2CH3), 1.56-1.80 (m, 4H, 2 CH2CH2CH3), 2.59 (t, J=7.5 Hz, 2H, CH2CH2CH3), 2.91 (br t, J=11.7 Hz, 2H, 2 SO2NCHax), 3.12-3.27 (m, 4H, NCH2CH2 and 2 SO2NCHeq), 3.58 (br d, J=11.7 Hz, 2H, 2 +HNCHax), 3.68-3.85 (m, 4H, CH2CH2OH and 2 +HNCHeq), 4.00 (s, 3H, NCH3), 4.15 (t, J=6.3 Hz, 2H, OCH2CH2CH3), 4.66 (br s, 1H, OH), 7.28 (s, 1H, H-2), 7.44 (d, J=9.0 Hz, 1H, H-3′), 7.89 (dd, J=9.0 Hz, 2.4 Hz, 1H, H-4′), 8.01 (d, J=2.4 Hz, 1H, H-6′), 10.85 (br s, 1H, NH+), 12.01 (br s, 1H, NH).
Example 42 Preparation of
5-(5-(4-(3-hydroxypropyl)piperazinylsulfonyl)-2-n-propoxyphenyl)-1-methyl-3-n-propyl-1,6-dihydro-7H-pyrrolo[4,3-d]pyrimidin-7-one (a compound of the formula (1) wherein R5=SO2NR6R7, R1=CH3, R2=H, R3=CH2CH2CH3, R4=CH2CH2CH3; NR6R7 is 4-(3-hydroxypropyl)piperazinyl)
The titled compound was prepared as described in Example 1 by using 5-(5-chlorosulfonyl-2-n-propoxyphenyl)-1-methyl-3-n-propyl-1,6-dihydro-7H-pyrrolo[4,3-d]pyrimidin-7-one and 1-(3-hydroxypropyl)piperazine in place of 5-(5-chlorosulfonyl-2-ethoxyphenyl)-1-methyl-3-n-propyl-1,6-dihydro-7H-pyrrolo[4,3-d]pyrimidin-7-one and 1-methylpiperazine.
yield: 94%
mp 162.5° C. dec (EtOAc/hexanes);
IR (neat) 3484, 3302 (NH and OH), 1669 (C═O), 1170 (SO2) cm−1;
1H NMR (CDCl3/TMS) δ 1.00 (t, J=7.5 Hz, 3H, CH2CH2CH3), 1.20 (t, J=7.5 Hz, 3H, OCH2CH2CH3), 1.64-1.80 (m, 4H, CH2CH2CH2OH and CH2CH2CH3), 1.99-2.11 (m, 2H, OCH2CH2CH3), 2.58-2.64 (m, 6H, NCH2CH2 and 2 NCH2), 2.71 (t, J=7.5 Hz, 2H, CH2CH2CH3), 3.08 (br s, 4H, 2 SO2NCH2), 3.71 (t, J=5.4 Hz, 2H, CH2CH2OH), 4.08 (s, 3H, NCH3), 4.26 (t, J=6.3 Hz, 2H, OCH2CH2CH3), 4.28 (br s, 1H, OH), 6.88 (s, 1H, H-2), 7.14 (d, J=8.7 Hz, 1H, H-3′), 7.77 (dd, J=8.7 Hz, 2.7 Hz, 1H, H-4′), 8.87 (d, J=2.7 Hz, 1H, H-6′), 10.69 (br s, 1H, NH); MS (FAB) m/z 532 (MH+).
Example 43 Preparation of
5-(5-(4-(3-hydroxypropyl)piperazinylsulfonyl)-2-n-propoxyphenyl)-1-methyl-3-n-propyl-1,6-dihydro-7H-pyrrolo[4,3-d]pyrimidin-7-one hydrochloride (a compound of the formula (1) wherein R5=SO2NR6R7, R1=CH3, R2=H, R3=CH2CH2CH3, R4=CH2CH2CH3; NR6R7 is 4-(3-hydroxypropyl)piperazinyl)
The titled compound was prepared as described in Example 2 by using 5-(5-(4-(3-hydroxypropyl)piperazinylsulfonyl)-2-n-propoxyphenyl)-1-methyl-3-n-propyl-1,6-dihydro-7H-pyrrolo[4,3-d]pyrimidin-7-one in place of 5-(2-ethoxy-5-(4-methylpiperazinylsulfonyl)phenyl)-1-methyl-3-n-propyl-1,6-dihydro-7H-pyrrolo[4,3-d]pyrimidin-7-one.
yield: 99%
mp 62.5° C. dec;
IR (neat) 3347, 3321 (NH and OH), 1689 (C═O), 1168 (SO2) cm−1;
1H NMR (DMSO-d6) δ 0.93 (t, J=7.5 Hz, 3H, CH2CH2CH3), 0.96 (t, J=7.5 Hz, 3H, OCH2CH2CH3), 1.57-1.87 (m, 6H, CH2CH2CH2OH and 2 CH2CH2CH3), 2.59 (t, J=7.5 Hz, 2H, CH2CH2CH3), 2.89 (br t, J=11.7 Hz, 2H, 2 SO2NCHax), 3.01-3.19 (m, 4H, NCH2CH2 and 2 SO2NCHeq), 3.44 (t, J=6.0 Hz, 2H, CH2CH2OH), 3.52 (br d, J=11.7 Hz, 2H, 2 +HNCHax), 3.79 (br d, J=11.7 Hz, 2H, 2 +HNCHeq), 4.00 (s, 3H, NCH3), 4.15 (t, J=6.6 Hz, 2H, OCH2CH2CH3), 4.71 (br s, 1H, OH), 7.29 (s, 1H, H-2), 7.44 (d, J=8.7 Hz, 1H, H-3′), 7.89 (dd, J=8.7 Hz, 2.4 Hz, 1H, H-4′), 8.02 (d, J=2.4 Hz, 1H, H-6′), 11.13 (br s, 1H, NH+), 12.05 (br s, 1H, NH).
……………………………
Synthesis from patent and some construction by me
you can synthesize as follows, A CHEMIST CAN PICK THIS UP, this is not available clearly anywhere
Chlorosulfonation of the methyl salicylate with ClSO3H in SOCl2 affords the Methyl 3-Chlorosulfonyl-6-hydroxybenzoate described below
THESE INTERMEDIATES FROM PATENT MAY HELP YOU
 methyl salicylate
 X=CL, R8=ME
- Methyl 3-Chlorosulfonyl-6-hydroxybenzoate
Example 1 EP1362858A1
-
To a cooled solution of SOCl2 (156 g, 1. 31 mol) and ClSO3H (460 g, 3.94 mol) at 0°C was added slowly methyl salicylate (200 g, 1.31 mol) for 30 minutes, and the mixture was stirred at room temperature for 20 hours. The reaction mixture was poured slowly into the ice (2 kg) and H2O (3 L) mixture, and the resulting white precipitates were collected by filtration. The filtered solid was washed with H2O (3 L), air-dried for 2 days and then dried under vacuum at 40°C for 2 days to afford the titled product (232 g, 93%) as a white solid.
mp 76.5-77.5 °C (toluene/hexanes);
IR (neat) 1699 (C=O) cm-1;
1H NMR (CDCl3/TMS) δ 3. 90 (s, 3 H, OCH3), 6. 93 (d, J= 8. 7 Hz, 1 H, H-3), 7. 70 (dd, J= 8. 7 Hz, 2. 4 Hz, 1 H, H-4), 8. 03 (d, J= 2. 4 Hz, 1 H, H-6).
- Methyl 3-Chlorosulfonyl-6-hydroxybenzoate
Example 2 EP1362858A1
-
 1-(2-hydroxyethyl)piperazine
 R8=ME, W=N, n=2
-
- Methyl 2-Hydroxy-5-[4-(2-hydroxyethyl)piperazin-1-ylsulfonyl]benzoate
-
To a mixture of 1-(2-hydroxyethyl)piperazine (27 mg, 0. 21 mmol) and K2CO3 (33 mg, 0. 24 mmol) in DMF (5 mL) was added methyl 3-chlorosulfonyl-6-hydroxybenzoate (50 mg, 0. 20 mmol), and the mixture was stirred at room temperature for 1 hour. The reaction mixture was washed with H2O (10 mL), and the aqueous layer was further extracted with 5% MeOH in CH2Cl2 (20 mL). The combined organic layer was dried (MgSO4), filtered, and the filtrate was evaporated to dryness under reduced pressure. The crude residue was purified by MPLC on silica gel (5% MeOH in CH2Cl2) to afford the titled compound (59 mg, 86%) as white solid.
mp 152 °C (dec) (CH2Cl2/ether);
IR (neat) 1685 (C=O) cm-1;
1H NMR (CDCl3/TMS) δ 2. 30 (br s, 1 H, CH2OH), 2. 63 (t, J = 5. 4 Hz, 2 H, NCH 2CH2O), 2. 70 (m, 4 H, 2 NCH2), 3. 12 (m, 4 H, 2 SO2NCH2), 3. 64 (t, J= 5. 4 Hz, 2 H, NCH2CH 2O), 4. 01 (s, 3 H, OCH3), 7. 12 (d, J= 8. 7 Hz, 1 H, H-3), 7. 81 (dd, J= 8. 7 Hz, 2. 4 Hz, 1 H, H-4), 8. 26 (d, J = 2. 4 Hz, 1 H, H-6), 11. 26 (br s, 1 H, OH);
MS (FAB) m/z 345 (MH+).
- Methyl 2-Hydroxy-5-[4-(2-hydroxyethyl)piperazin-1-ylsulfonyl]benzoate
Example 3 EP1362858A1
Methyl 3-[4-(2-Hydroxyethyl)piperazin-1-ylsulfonyl]-6-n-propoxybenzoate
-
To a mixture of methyl 2-hydroxy-5-(4-(2-hydroxyethyl)piperazin-1-ylsulfonyl)benzoate (800 mg, 2. 32 mmol) and K2CO3 (482 mg, 3. 49 mmol) in DMF (5 mL) was added 1-bromopropane (253 µL, 2.79 mmol), and the mixture was stirred at 60°C overnight. The reaction mixture was evaporated to dryness under reduced pressure, washed with H2O (10 mL), and the aqueous layer was further extracted with CH2Cl2 (50 mL x 2). The combined organic layer was dried (MgSO4), filtered, and the filtrate was evaporated to dryness under reduced pressure. The crude residue was purified by MPLC on silica gel (3% MeOH in CHCl3) to afford the titled compound (309 mg, 80%) as a white solid.
mp 88-89 °C (EtOAc/hexanes);
IR (neat) 3242 (OH), 1741 (C=O) cm-1;
1H NMR (CDCl3/TMS) δ 1. 09 (t, J = 7. 5 Hz, 3 H, OCH2CH2CH 3), 1. 84-1. 95 (m, 2 H, OCH2CH 2CH3), 2. 23 (br s, 1 H, CH2OH), 2. 54 (t, J= 5. 4 Hz, 2 H, NCH 2CH2O), 2. 60 (m, 4 H, 2 NCH2), 3. 04 (m, 4 H, 2 SO2NCH2), 3. 58 (t,J = 5. 4 Hz, 2 H, NCH2CH 2O), 3. 91 (s, 3 H, OCH3), 4. 08 (t, J= 6. 6 Hz, 2 H, OCH 2CH2CH3), 7. 07 (d, J = 9. 0 Hz, 1 H, H-3), 7. 82 (dd, J = 9. 0 Hz, 2. 4 Hz, 1 H, H-4), 8. 15 (d, J = 2. 4 Hz, 1 H, H-6);
MS (FAB) m/z 387 (MH+).
- FURTHER INFO OTHER THAN ABOVE PATENT
- HYDROLYSE Methyl 3-[4-(2-Hydroxyethyl)piperazin-1-ylsulfonyl]-6-n-propoxybenzoate TO -COOLi SALT using LiOH
- CONDENSE WITH 3-amino-1-ethyl-4-propyl-1H-pyrrole-2-carboxamide USING HOBt AND DMAP/ PYRIDINE

9……….Methyl 3-[4-(2-Hydroxyethyl)piperazin-1-ylsulfonyl]-6-n-propoxybenzoate R8= ME, R4=PROPYL, W=N, n=2
10……….3-amino-1-ethyl-4-propyl-1H-pyrrole-2-carboxamide R1=ETHYL, R2=H, R3=PROPYL, IN ABOVE
YOU WILL GET A COMPD

R1 ETHYL
R2=H
R3= PROPYL
R4 = PROPYL
W=N
n=2
IS MIRODENAFIL precursor ie n-1 compund
- CYCLIZE THIS WITH BuOK/tBuOH AND USE ACID TO GET FINAL PRODUCT MIRODENAFIL
- A cyclization reaction is generally carried out by heating at an elevated temperature, for example 50-150° C., in the presence of an acid or a base in a suitable solvent such as an aqueous C1-C4 alkanol, water, a halogenated hydrocarbon, or acetonitrile. Thus, for example, the cyclization may be affected by treatment of a compound with an inorganic or organic base such as sodium hydroxide, potassium carbonate or potassium tert-butoxide, in an alcoholic aqueous medium, preferably potassium tert-butoxide in tert-butanol at 60° C. to reflux temperature.
SYNTHESIS OF 1-(2-hydroxyethyl)piperazine needed for MIRODENAFIL SYNTHESIS
Compounds of the formula (29) can be prepared from the compounds of the formula (30):

wherein X and P are as previously defined.
note X=N ATOM, n = 2
…………………………….
 MIRODENAFIL
Two methods were published for the determination of mirodenafil in biological fluids. Choi et al. (2009) describe an isocratic reversed-phase liquid chromatographic method for simultaneous analysis of mirodenafil and its two main metabolites, SK3541 and SK3544, in rat plasma, urine, and tissue homogenates. The authors used a simple deproteinization procedure for sample preparation, and the compounds were separated on a C18 column (250 mm x 4.6 mm, i.d.; 5 µm particle size; Shiseido, Tokyo, Japan). The mobile phase was constituted with 0.02 M ammonium acetate buffer (pH 6):acetonitrile (52:48, v/v) at a flow rate of 1.4 mL/min. UV detection was at 254 nm.
Lee et al. (2009) developed a study with the proposed method to determine sildenafil and mirodenafil in the plasma and corpus cavernosum tissue of rats using LC–MS/MS. A CapcellPak phenyl column (2.1mm x 150 mm, 5µm) maintained constant at 40 ºC was used for the separation. The mobile phase consisted of 90% acetonitrile in 5 mM ammonium formate (pH 6.0). A gradient program was used for the LC separation with a flow rate of 0.2 mL/min.
References
- Paick JS, Ahn TY, Choi HK, Chung WS, Kim JJ, Kim SC, Kim SW, Lee SW, Min KS, Moon KH, Park JK, Park K, Park NC, Suh JK, Yang DY, Jung HG (November 2008). “Efficacy and safety of mirodenafil, a new oral phosphodiesterase type 5 inhibitor, for treatment of erectile dysfunction”. The Journal of Sexual Medicine 5 (11): 2672–80. doi:10.1111/j.1743-6109.2008.00945.x. PMID 18638004.
- Kim BH, Yi S, Kim J, Lim KS, Kim KP, Lee B, Shin SG, Jang IJ, Yu KS (June 2009). “Influence of alcohol on the hemodynamic effects and pharmacokinetic properties of mirodenafil: a single-dose, randomized-sequence, open-label, crossover study in healthy male volunteers in Korea”.Clinical Therapeutics 31 (6): 1234–43. doi:10.1016/j.clinthera.2009.06.008. PMID 19695390.
- Shin KH, Kim BH, Kim TE, Kim JW, Yi S, Yoon SH, Cho JY, Shin SG, Jang IJ, Yu KS (December 2009). “The effects of ketoconazole and rifampicin on the pharmacokinetics of mirodenafil in healthy Korean male volunteers: an open-label, one-sequence, three-period, three-treatment crossover study”.Clinical Therapeutics 31 (12): 3009–20. doi:10.1016/j.clinthera.2009.12.012. PMID 20110038.
- Synthesis of 5-ethyl-2-[5-[4-(2-hydroxyethyl)piperazin-1-ylsulfonyl]-2-n-propoxyphenyl]-7-n-propyl-3,5-dihydro-4H-pyrrolo[3,2-d]-[2-14C]pyrimidin-4-one·2 HCl (14C-SK3530·2 HCl)J Label Compd Radiopharm 2006, 49(13): 1141
- More information about mirodenafil can be found at Paick J S et al., (2008) The Journal of Sexual Medicine, 5 (11): 2672-80.
- PDE-5 inhibitor that came into the market recently (Choi et al., 2009; Lee et al., 2009).not currently approved for use in the United States but clinical trials are being conducted.
- Crystal forms of SK-3530.
Song HO, Sohn YT.Arch Pharm Res. 2010 Dec;33(12):2033-6. doi: 10.1007/s12272-010-1220-3. Epub 2010 Dec 30. - Looking to the future for erectile dysfunction therapies.Hatzimouratidis K, Hatzichristou DG.Drugs. 2008;68(2):231-50. Review.
-
- Paick JS, Ahn TY, Choi HK, Chung WS, Kim JJ, Kim SC, Kim SW, Lee SW, Min KS, Moon KH, Park JK, Park K, Park NC, Suh JK, Yang DY, Jung HG (November 2008). “Efficacy and safety of mirodenafil, a new oral phosphodiesterase type 5 inhibitor, for treatment of erectile dysfunction”. The Journal of Sexual Medicine 5 (11): 2672–80. doi:10.1111/j.1743-6109.2008.00945.x. PMID 18638004.
- Kim BH, Yi S, Kim J, Lim KS, Kim KP, Lee B, Shin SG, Jang IJ, Yu KS (June 2009). “Influence of alcohol on the hemodynamic effects and pharmacokinetic properties of mirodenafil: a single-dose, randomized-sequence, open-label, crossover study in healthy male volunteers in Korea”. Clinical Therapeutics 31 (6): 1234–43. doi:10.1016/j.clinthera.2009.06.008. PMID 19695390.
- Shin KH, Kim BH, Kim TE, Kim JW, Yi S, Yoon SH, Cho JY, Shin SG, Jang IJ, Yu KS (December 2009). “The effects of ketoconazole and rifampicin on the pharmacokinetics of mirodenafil in healthy Korean male volunteers: an open-label, one-sequence, three-period, three-treatment crossover study”. Clinical Therapeutics 31 (12): 3009–20. doi:10.1016/j.clinthera.2009.12.012. PMID 20110038.
- Matheny, C., et al., Drug Metab. Dispos., 32, 1008 (2004)
Gupta, M., et al., J. Clin. Pharmacol., 45, 987 (2005)
Ek, M., et al., Biochem. Pharmacol., 74, 496 (2007)
Lee, H., et al., Xenobiotica, 38, 21 (2008)

| WO2006018088A1 * | Jul 15, 2005 | Feb 23, 2006 | Switch Biotech Ag | Use of a pde 5 inhibitor for treating and preventing hypopigmentary disorders |
| KR20010083637A * | Title not available |
| US6962911 * | Feb 15, 2001 | Nov 8, 2005 | Sk Chemicals Co., Ltd. | Pyrrolopyrimidinone derivatives, process of preparation and use |
| US20100069632 * | Jul 3, 2007 | Mar 18, 2010 | Sk Chemicals Co., Ltd | Salts of pyrrolopyrimidinone derivatives and process for preparing the same |
| EP2038282A1 * | Jul 3, 2007 | Mar 25, 2009 | SK Chemicals, Co., Ltd. | Salts of pyrrolopyrimidinone derivatives and process for preparing the same |


DASANTAFIL
 Dasantafil

569351-91-3 CAS NO
405214-79-1 (racemate)
THERAPEUTIC CLAIM treatment of erectile dysfunction (phosphodiesterase (PDE) 5 isoenzyme inhibitor)
read all at
ALL ABOUT DRUGS
CLICK BELOW
http://www.allfordrugs.com/2014/01/29/dasantafil-for-treatment-of-erectile-dysfunction/
Gisadenafil for erectile dysfunction
GISEDENAFIL
GISEDENAFIL BESYLATE
334826-98-1 free form
334827-98-4 (as besylate)
- UK 369003
- UK-369,003
- UK0369,003
- UNII-S6G4R7DI1C
THERAPEUTIC CLAIM Treatment of lower urinary tract
symptoms associated with BPH
LEARN SPECTROSCOPY USING GISADENAFIL INTERMEDIATES
CHEMICAL NAMES FREE FORM
1. ……..7H-Pyrazolo[4,3-d]pyrimidin-7-one, 5-[2-ethoxy-5-[(4-ethyl-1-
piperazinyl)sulfonyl]-3-pyridinyl]-3-ethyl-2,6-dihydro-2-(2-methoxyethyl)-
2. …….5-{2-ethoxy-5-[(4-ethylpiperazin-1-yl)sulfonyl]pyridin-3-yl}-3-ethyl-2-(2-
methoxyethyl)-2,6-dihydro-7H-pyrazolo[4,3-d]pyrimidin-7-one
3………1-(6-Ethoxy-5-[3-ethyl]-6,7-dihydro-2-(2-methoxyethyl)-7-oxo-2H-pyrazole[4,3-d]pyrimidin-5-yl]-3-pyridylsulfonyl)-4-ethylpiperazine
MOLECULAR FORMULA C23H33N7O5S
MOLECULAR WEIGHT 519.6
CODE DESIGNATION UK-369,003
CAS REGISTRY NUMBER 334826-98-1
5-[2-Ethoxy-5-(4-ethylpiperazin-1-ylsulfonyl)pyridin-3-yl]-3-ethyl-2-(2-methoxyethyl)-6,7-dihydro-2H-pyrazolo[4,3-d]pyrimidin-7-one
Phosphodiesterase PDE5A Inhibitors , Treatment of Erectile Dysfunction
Pfizer (Originator)
UK-369003 is a phosphodiesterase V (PDE V) inhibitor which had been under development for the treatment of erectile dysfunction, pulmonary hypertension and for the treatment of lower urinary tract symptoms, but no recent development has been reported for these indications. Trials for the treatment of benign prostatic hyperplasia were discontinued.
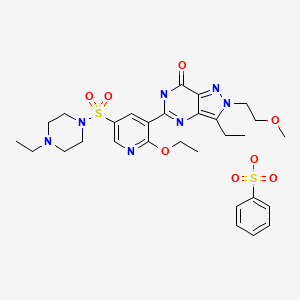
D09622, 334827-98-4
M.Wt:677.79
5-(2-ethoxy-5-(4-ethylpiperazin-1-ylsulfonyl)pyridin-3-yl)-3-ethyl-2-(2-methoxyethyl)-2H-pyrazolo[4,3-d]pyrimidin-7(6H)-one benzenesulfonate
1-[[6-Ethoxy-5-[3-ethyl-4,7-dihydro-2-(2-methoxyethyl)-7-oxo-2H-pyrazolo[4,3-d]pyrimidin-5-yl]-3-pyridinyl]sulfonyl]-4-ethylpiperazine Monobenzenesulfonate
Formula:C23H33N7O5S.C6H6O3S
| Certificate of Analysis |
|
| Biological Activity:Potent and selective PDE5 inhibitor (IC50: 1.23 nM) with improved selectivity over PDE6(PDE5/6 selectivity value 117 and >3000-fold selectivity over other PDEs).Gisadenafil has the potential for oral bioavailability and dose-proportional pharmacokinetics. Close analogue of Sildenafil (Viagra; Axon 2046) |
Gisadenafil besylate is a PDE5 inhibitor. Inhibition of PDE5 prevents the breakdown of cyclic phosphodiester secondary messenger molecules. This has the effect of prolonging and enhancing signal transduction.
CLINICAL TRIALS
http://clinicaltrials.gov/search/intervention=UK-369,003
………………………….
PAPERS
Bioorganic and Medicinal Chemistry, 2012 , vol. 20, 1 p. 498 – 509
http://www.sciencedirect.com/science/article/pii/S0968089611008303


Scheme 1.
Reagents and conditions: (i) 1-(3-dimethylaminopropyl)-3-ethyl-carbodiimide hydrochloride, hydroxybenzotriazole, di-isopropylethylamine, THF, 20 °C, 20 h; (ii) caesium carbonate, alkyl mesylate or alkyl chloride, DMF, 20 °C, 20 h; (iii) KHMDS, R1OH, 120 °C, 20 h.

Scheme 2.
Reagents and conditions: (i) KHMDS, nBuOH, 120–130 °C, pressure vessel (ii) TFA, CH2Cl2; (iii) methanesulphonyl chloride, NEt3, CH2Cl2; (iv) HOAc, NaCNBH3, CH2O (v) KHMDS, nBuOH, reflux.

Scheme 3.
Reagents and conditions: (i) caesium carbonate, RCl, DMF; (ii) 50 psi H2, 10% Pd/C (iii) 1-(3-dimethylaminopropyl)-3-ethyl-carbodiimide hydrochloride, HOBT, di-isopropylethylamine, THF, 20 °C, 20 h; (iv) KHMDS, ethanol, 120 °C, pressure vessel; (v) TFA, CH2Cl2; (vi) CH2O, HOAc, NaCNBH3; (vii) R1OH, KHMDS, 120 °C.

Scheme 4.
Reagents and conditions: (i) NaNO2, HCl, H2O; (ii) TFAA, Et2O; (iii) ethyl propynoate, xylene, reflux, 2 h; (iv) NaOH, H2O, dioxan; (v) HNO3/H2SO4, 40–55 °C; (vi) (COCl)2, CH2Cl2, DMF; (vii) NH3, THF; (viii) 10% Pd/C, EtOH, 60 psi H2, 20 °C, 14 h; (ix) acid chloride of 3, NEt3, CH2Cl2; (x) KHMDS, EtOH, 130 °C, 14 h, pressure vessel; (xi) methoxyethanol, KHMDS, reflux, 14 h.
……………………………
PAPERS
http://pubs.acs.org/doi/abs/10.1021/op0300241

………………………….

PAPERS

UK-369,003 was nominated for development as the lead candidate for treatment of benign prostatic hyperplasia (BPH). The free base was found to be moderately crystalline with a melting point of 168 °C. Solubility of the free base at physiological pH was found to be poor hence necessitating a comprehensive screen for a suitable salt form of the API. Benzenesulfonic acid was found to form the most suitable counterion for the API with a melting point of 248 °C and satisfied all our requirements for primary and secondary processing. The process for the formation of the benzenesulfonic acid salt involved the use of water/methyl ethyl ketone (4% water by volume) as the reaction medium. The water level at 4% ensured an optimum balance between product quality (purging of impurities) and the reaction yield. The cyclisation reaction (step 2/Scheme 01) involves the use of ethanol as the reaction media. Any residual amount of ethanol in the isolated step 2 product was therefore considered to be a considerable risk factor in the potential formation of ethyl besylate during the final step processing (step 3/Scheme 01).

aCDI = carbonyl diimidazole; MEK = methyl ethyl ketone; EtOAc = ethyl acetate; KOtBu = potassium tertiary butoxide; EtOH = ethanol.
……………………
SYNTHESIS
Compound 1E is also known as 5-[2-Ethoxy-5-(4-ethylpiperazin-1-ylsulphonyl)pyridin-3-yl]-3-ethyl-2-[2-methoxyethyl]-2,6-dihydro-7H-pyrazolo[4,3-d]pyrimidin-7-one, or alternatively as 1-{6-ethoxy-5-[3-ethyl-6,7-dihydro-2-(2-methoxyethyl)-7-oxo-2H-pyrazolo[4,3-d]pyrimidin-5-yl]-3-pyridyl sulphonyl}-4-ethylpiperazine (the compound of Example 103 of WO 01/27113 and exemplified hereinafter as Example 1).
Preparation 1
2,2-dimethoxybutane:
Methyl ethyl ketone (672 mL) was charged to a 2 L round bottomed flask and stirred at room temperature before being treated with, trimethylorthoformate (763 mL) and para-toluenesulphonic acid (6.65 g, 0.5 mol %). Over a 15 min period the internal temperature rose to 46° C., so the reaction was cooled to 0° C. for 30 min. The reaction was then stirred at room temperature for 2 h. The reaction was then neutralised by pouring onto sodium carbonate (ca. 750 g) with constant stirring. The resultant slurry was filtered under vacuum and the resultant filtrate was distilled at atmospheric pressure. The fraction boiling in the range 118° C.-124° C. was collected as a colourless liquid, 582 g, 70%.
1H NMR (CDCl3): δ=0.88 (3H, t), 1.24 (3H, s), 1.61 (2H, q), 3.17 (6H, s).
Example 1 N-[3-Carbamoyl-5-ethyl-1-(2-methoxyethyl)-1H-pyrazol-4-yl]-2-ethoxy-5-(4-ethyl-1-piperazinyl sulfonyl) nicotinamide
(a) Ethyl 3-ethyl-1H-pyrazole-5-carboxylate (IIA) from (IlI) and (V)

To a stirred solution of 2,2-dimethoxybutane (10 g, 84.7 mMol) in CH2Cl2 (50 mL) under a nitrogen atmosphere at 0° C. was added pyridine (13.7 mL, 169.5 mMol). The reaction mixture was maintained at 0° C. and a solution of trichloroacetyl chloride (18.9 mL, 169.5 mMol) in CH2Cl2 (35 mL) was added over 1 hour with constant stirring. The yellow-orange solution begins to precipitate a white solid as the reaction progresses. The reaction mixture is allowed to warm to room temperature over 20 h. The reaction mixture was diluted with ethanol (150 mL) and re-cooled to 0° C. before treatment with hydrazine hydrate (8.2 mL, 169.5 mMol) as a solution in ethanol (35 mL) over 30 min. The reaction was heated to 50° C. and solvent was distilled at atmospheric pressure. The temperature was increased until the head temperature reached 78° C. Reflux was maintained for a further 2 h, before cooling to room temperature. The reaction mixture was diluted with water (250 mL) and ethanol was removed by evaporation at reduced pressure. The resultant mixture was extracted with CH2Cl2 (3×200 mL). The combined organics were dried (MgSO4), filtered and evaporated at reduced pressure to afford the title compound as a brown oil, 12.05 g, 85%.
1H NMR (300 MHz, CDCl3): δ=1.20 (3H, t), 1.28 (3H, t), 2.67 (2H, q), 4.29 (2H, q), 6.55 (1H, s), 12.56 (1H, s).
LRMS m/z=167.1 [M-H]+, C8H12N2O2 requires 168.2.
(b) Ethyl 3-ethyl-1H-pyrazole-5-carboxylic acid (IIA) from (IIA) via route 1

Aqueous sodium hydroxide solution (10M; 100 ml, 1.0 mol) was added dropwise to a stirred suspension of the title compound of Example (a) (66.0 g, 0.39 mol) in methanol and the resulting solution heated under reflux for 4 hours. The cool reaction mixture was concentrated under reduced pressure to ca. 200 ml, diluted with water (200 ml) and this mixture washed with toluene (3×100 ml). The resulting aqueous phase was acidified with concentrated hydrochloric acid to pH 4 and the white precipitate collected and dried by suction to provide the title compound (34.1 g). δ (DMSOd6): 1.13 (3H,t), 2.56 (2H,q), 6.42 (1H,s).
(c) 4-Nitro-3-n-propyl-1H-pyrazole-5-carboxylic acid
Fuming sulphuric acid (17.8 ml) was added dropwise to stirred, ice-cooled fuming nitric acid (16.0 ml), the resulting solution heated to 50° C., then 3-n-propyl-1H-pyrazole-5-carboxylic acid (Chem. Pharm. Bull., 1984, 32,1568; 16.4 g, 0.106 mol) added portionwise over 30 minutes whilst maintaining the reaction temperature below 60° C. The resulting solution was heated for 18 hours at 60° C., allowed to cool, then poured onto ice. The white precipitate was collected, washed with water and dried by suction to yield the title compound (15.4 g), m.p. 170-172° C. Found: C, 42.35; H, 4.56; N, 21.07. C7H9N3O4requires C, 42.21; H, 4.55; N, 21.10%. δ (DMSOd6): 0.90 (3H,t), 1.64 (2H,m), 2.83 (2H,m), 14.00 (1 H,s).
(d) 3-Ethyl-4-nitro-1H-pyrazole-5-carboxylic acid (IIA) to (AA) via route 2

Obtained from the title compound of Example (b), by analogy with the process of Example (c), as a brown solid (64%). δ (DMSOd6): 1.18 (3H,t), 2.84 (2H,m), 13.72 (1 H,s).
(e) 4-Nitro-3-n-propyl-1H-pyrazole-5-carboxamide
A solution of the title compound of Example (c) (15.4 g, 0.077 mol) in thionyl chloride (75 ml) was heated under reflux for 3 hours and then the cool reaction mixture evaporated under reduced pressure. The residue was azeotroped with tetrahydrofuran (2×50 ml) and subsequently suspended in tetrahydrofuran (50 ml), then the stirred suspension ice-cooled and treated with gaseous ammonia for 1 hour. Water (50 ml) was added and the resulting mixture evaporated under reduced pressure to give a solid which, after trituration with water and drying by suction, furnished the title compound (14.3 g).
m.p. 197-199° C. Found: C, 42.35; H, 5.07; N, 28.38. C7H10N4O3 requires C, 42.42; H, 5.09; N, 28.27%. δ (DMSOd6): 0.90 (3H,t), 1.68 (2H,m), 2.86 (2H,t), 7.68 (1 H,s), 8.00 (1 H,s).
(f) 3-Ethyl-4-nitro-1H-pyrazole-5-carboxamide BA from AA via route 3

Obtained from the title compound of Example (d), by analogy with Example (e), as a white solid (90%). δ (DMSOd6): 1.17 (3H,t), 2.87 (2H,m), 7.40 (1H,s), 7.60 (1H,s), 7.90 (1H,s). LRMS: m/z 185 (M+l)+.
(g)(i) 5-Ethyl-1-(2-methoxyethyl)-4-nitro-1H-pyrazole-3-carboxamide CA from BA via route 4

A mixture of 3-ethyl-4-nitro-1H-pyrazole-5-carboxamide (2.5 kg, 13.6 Mol), sodium carbonate (1.8 Kg, 17.0 Mol) and 2-bromoethyl methyl ether (1.98 kg, 14.2 Mol) in THF (22.5 L) and water (2.5 L) was heated under reflux and stirred for 20 hours. The mixture was cooled to ambient temperature and CH2Cl2 (67.5 L) and water (22.5 L) were added. The resultant organic and aqueous layers were separated. The aqueous phase was extracted with CH2Cl2 (22.5 L) and the combined organic solution was distilled under atmospheric pressure and replaced with ethyl acetate (33 L) to a final volume of 17 L. The cooled mixture was granulated at ambient temperature for 2 hours, filtered and washed with ethyl acetate (2.5 L). This afforded 5-ethyl-1-(2-methoxyethyl)-4-nitro-1H-pyrazole-3-carboxamide as a white crystalline solid, 2.10 kg, 57%. m.p.=140° C. Found: C, 44.46; H, 5.79; N, 23.01. C9H14N4O4 requires C, 44.63; H, 5.79; N, 23.14%.
δ (CDCl3): 1.18 (3H, t), 2.98 (2H, q), 3.22 (3H, s), 3.77 (2H, t), 4.28 (2H, q), 6.03 (1H, s), 7.36 (1H, s).
LRMS: m/z=243 (M+1)+
(g)(ii) 5-Ethyl-1-(2-methoxyethyl)-4-nitro-1H-pyrazole-3-carboxamide.
A mixture of 3-ethyl-4-nitro-1H-pyrazole-5-carboxamide (25 g, 0.136 Mol), sodium carbonate (18 g, 0.17 Mol) and sodium iodide (20.4 g, 0.136 Mol) were suspended in ethyl methyl ketone (125 mL) at room temperature. 2-bromoethyl methyl ether (12.8 mL, 0.142 Mol) was added and the mixture was heated to reflux and stirred for 70 hours. The mixture was cooled to ambient temperature and water (250 mL) was added. The resultant slurry was warmed to reflux and held at that temperature for 30 min before cooling to room temperature. The resultant precipitate was granulated at room temperature for 3 h, filtered and vacuum dried to afford 5-ethyl-1-(2-methoxyethyl)-4-nitro-1H-pyrazole-3-carboxamide as a yellow crystalline solid 24.3 g, 74%. Data as reported for Example (g)(i).
(h) 4-Amino-5-ethyl-1-(2-methoxyethyl)-1H-pyrazole-3-carboxamide (IA) from CA via route 5

A mixture of 5-ethyl-1-(2-methoxyethyl)-4-nitro-1H-pyrazole-3-carboxamide (20 g, 82.6 mMol) and 5% Pd/C (1 g) in methanol (200 mL) was pressurised at 50psi/25° C. in a sealed vessel and stirred for 15 hours. At the end of the reaction the mixture was filtered through arbocel and the filter cake was washed with methanol. The methanolic solution was distilled at atmospheric pressure and replaced with ethyl acetate to a final volume of 100 mL. The cooled mixture was granulated at ambient temperature for 2 h filtered and washed with ethyl acetate (20 mL) to afford 4-amino-5-ethyl-1-(2-methoxyethyl)-1H-pyrazole-3-carboxamide as a white crystalline solid, 15 g, 88%. m.p.=131° C. Found: C, 50.75; H, 7.62; N, 26.38. C9H16N4O2 requires C, 50.94; H, 7.55; N, 26.42%. δ (CDCl3): 1.20 (3H, t), 2.63 (2H, q), 3.32 (3H, s), 3.74 (2H, t), 3.95 (2H, s), 4.15 (2H, t), 5.27 (1H, s), 6.59 (1H, s).
LRMS: m/z=213 (M+1)+
(i) N-[3-Carbamoyl-5-ethyl-1-(2-methoxyethyl)-1H-pyrazol-4-yl]-2-ethoxy-5-(4-ethyl-1-piperazinyl sulfonyl) nicotinamide.

2-ethoxy-5-(4-ethyl-1-piperazinylsulfonyl)nicotinic acid (2.31 kg, 6.73 Mol) was suspended in ethyl acetate (16.2 L) and 1,1-carbonyldimidazole (1.09 kg, 6.73 Mol) was added at room temperature. The reaction mixture was heated at 45° C. for 40 minutes and then the reaction was stirred for a further 40 minutes at reflux. After cooling to ambient temperature 4-amino-5-ethyl-1-(2-methoxyethyl)-1H-pyrazole-3-carboxamide (1.5 kg, 7.06 Mol) was added to the cooled mixture, and the reaction stirred for a further 15 hours under reflux. The mixture was cooled filtered and the filter cake was washed with 90% water/10% ethyl acetate, (2 mL /g) to afford N-[3-carbamoyl-5-ethyl-1-(2-methoxyethyl)-1H-pyrazol-4-yl}-2-ethoxy-5-(4-ethyl-1-piperazinyl sulfonyl) nicotinamide as an off white crystalline solid, 3.16 kg, 88%. m.p.=156° C. Found: C, 51.33; H, 6.56; N, 18.36. C23H35N7O6S requires C, 51.40; H, 6.53; N, 18.25%.
δ (CDCl3): 1.04 (3H, t), 1.22 (3H, t), 1.60 (3H, t), 2.44 (2H, q), 2.54 (4H, m), 2.96 (2H, q), 3.12 (4H, m), 3.36 (3H, s), 3.81 (2H, t), 4.27 (2H, t), 4.80(2H, q), 5.35(1H, s), 6.68 (1H, s), 8.66 (1H, d), 8.86 (1H, d), 10.51 (1H, s).
LRMS: m/z=539 (M+1)+
(i) 1-(6-Ethoxy-5-[3-ethyll-6,7-dihydro-2-(2-methoxyethyl)-7-oxo-2H-pyrazole[4,3-d]pyrimidin-5-yl]-3-pyridylsulfonyl)-4-ethylpiperazine•ethyl acetate solvate.

GISADENAFIL
A mixture of N-[3-carbamoyl-5-ethyl-1-(2-methoxyethyl)-1H-pyrazol-4-yl}-2-ethoxy-5-(4-ethyl-1-piperazinyl sulfonyl) nicotinamide (1.18 kg, 2.2 Mol), potassium tert-butoxide (500 g, 4.4 moles) and ethyl acetate (193 g) in ethanol (11.8 L) was heated at 120° C. for 20 hours. The reaction mixture was then concentrated under reduced pressure, in total approx. 10 L of solvent were distilled. To the residue water (2.9 L) was added and the mixture stirred at room temperature while aqueous HCl was added until pH 7.5 was obtained. Ethyl acetate (7.5 L) was added and the two phase mixture was warmed to 55° C. The organic phase was separated and the aqueous phase was extracted with further ethyl acetate (3.0 L). The combined organic phases were distilled at atmospheric pressure to a final volume of 4 L. The precipitated solids were granulated at 5° C. for 1 h, filtered and washed with ethyl acetate (1.2 L) and dried under vacuum. This afforded 1-(6-Ethoxy-5-[3-ethyl]-6,7-dihydro-2-(2-methoxyethyl)-7-oxo-2H-pyrazole[4,3-d]pyrimidin-5-yl]-3-pyridylsulfonyl)-4-ethylpiperazine as a light yellow crystalline solid, 877 g, 78%. m.p.=157° C. Found: C, 52.65; H, 6.46; N, 17.76. C23H33N705S. 0.2 C2H5CO2CH3 requires C, 53.21; H, 6.49; N, 18.25%.
δ (CDCl3): 1.07 (3H, t), 1.42 (3H, t), 1.61 (3H, t), 2.44 (2H, q), 2.57 (4H, m), 3.08 (2H, q), 3.15 (4H, m), 3.32 (3H, s), 3.92 (2H, q), 4.48 (2H, q), 4.77 (2H, q), 8.65 (1H, d), 9.06 (1H, d). The spectrum also has signals that correspond to a solvate with ethyl acetate.
LRMS: m/z=520 (M+1)+
……………..
Example 102
1-(6-Ethoxy-5-f3-ethyll-6,7-dihvdro-2-(2-methoxyethvn-7-oxo-2r7-pyrazoler4.3- cf1pyrimidin-5-vn-3-pyridylsulfonyl)-4-ethylpiperazine»ethyl acetate solvate.

To prepare the compound of Example 8 a mixture of Λ/-[3-carbamoyl-5-ethyl- 1 -(2-methoxyethyl)-1 /-/-pyrazol-4-yl}-2-ethoxy-5-(4-ethyl-1 -piperazinyl sulfonyl) nicotinamide (1.18 kg, 2.2 Mol), potassium tert-butoxide (500 g, 4.4 moles) and ethyl acetate (193 g) in ethanol (11.8 L) was heated at 120°C for 20 hours. The reaction mixture was then concentrated under reduced pressure, in total approx. 10 L of solvent were distilled. To the residue water (2.9 L) was added and the mixture stirred at room temperature while aqueous HCl was added until pH 7.5 was obtained. Ethyl acetate (7.5 L) was added and the two phase mixture was warmed to 55°C. The organic phase was separated and the aqueous phase was extracted with further ethyl acetate (3.0 L). The combined organic phases were distilled at atmospheric pressure to a final volume of 4L. The precipitated solids were granulated at 5°C for 1 h, filtered and washed with ethyl acetate (1.2 L) and dried under vacuum. This afforded 1 -(6-Ethoxy-5-[3-ethyl]-6,7-dihydro-2-(2-methoxyethyl)-7-oxo- 2H-pyrazole[4,3-o pyrimidin-5-yl]-3-pyridylsulfonyl)-4-ethylpiperazine as a light yellow crystalline solid, 877 g, 78%. m.p. = 157°C. Found: C, 52.65; H, 6.46; N, 17.76. C23H33N705S. 0.2 C2H5C02CH3 requires C, 53.21 ; H, 6.49; N, 18.25%.
δ(CDCI3): 1.07 (3H, t), 1.42 (3H, t), 1.61 (3H, t), 2.44 (2H, q), 2.57 (4H, m), 3.08 (2H, q), 3.15 (4H, m), 3.32 (3H, s), 3.92 (2H, q), 4.48 (2H, q), 4.77 (2H, q), 8.65 (1 H, d), 9.06 (1 H, d). The spectrum also has signals that correspond to a solvate with ethyl acetate.
LRMS: m/z = 520 (M+1)+
Example 103
1-(6-ethoxy-5-r3-ethyl-6.7-dihvdro-2-(2-methoxyethvn-7-oxo-2H-pyrazolor4.3- dlpyrimidin-5-vn-3-pyridylsulfonyl)-4-ethylpiperazine

GISADENAFIL
10g (0.019 mol) of the compound of Example 8 and Example 102, 1-{6- ethoxy-5-[3-ethyl-6,7-dihydro-2-(2-methoxyethyl)-7-oxo-2H-pyrazolo[4,3- d]pyrimidin-5-yl]-3-pyridylsulfonyl}-4-ethylpiperazine ethyl acetate solvate, was charged followed by 12ml/g (120mls) of 16% water in ethyl alcohol. The slurry was heated to reflux to yield a solution and 6ml/g (60mls) distilled off at atmospheric pressure. The solution was then cooled to room temperature with crystallisation occurring at 40°C. The slurry was then cooled to 5-10°C and granulated for 30 minutes following which it was filtered and washed with 2ml/g ethyl alcohol (20 mis). The damp solid was dried in vacuo overnight at 55-60 °C to yield a white crystalline solid. (Yield 7.6g, 76%). Melting Point 162- 165°C.
δ (CDCI3): 1.05 (3H,t), 1.42 (3H,t), 1.58 (3H,t), 2.43 (2H,q), 2.57 (4H,t), 3.09 (2H, t), 3.15 (4H,t), 3.30 (3H,s), 3.93 (2H,t), 4.48 (2H,t), 4.90 (2H,q), 8.65 (1 H,d), 9.05 (1 H,d), 10.65 (1 H,s).
In the process of Example 103, water and pharmaceutically acceptable alcohols such as methanol, ethanol, propanol, butanol and mixtures thereof can be used to prepare the compound of Examples 8 and 102.
BESYLATE SALT
Example 104 1-(6-ethoxy-5-r3-ethyl-6,7-dihvdro-2-(2-methoxyethyl)-7-oxo-2H-pyrazolor4.3- d]pyrimidin-5-yl]-3-pyridylsulfonyl)-4-ethylpiperazine benzene-sulfonate salt.

170g (0.33 mol) of the compound of Example 103, 1-{6-ethoxy-5-[3-ethyl-6,7- dihydro-2-(2-methoxyethyl)-7-oxo-2H-pyrazolo[4,3- d]pyrimidin-5-yl]-3- pyridylsulfonyl}-4-ethylpiperazine, was charged followed by a water/ 2- butanone (4% v/v) at 10 ml/g (1.7 litres) and warmed to reflux. 53g (0.33 mol) of benzene sulfonic acid dissolved in water (23mls, resulting in 70 % w/w solution) was added to the refluxing solution over 30 minutes.5.3ml/g (0.9 litres) of 2-butanone were striped and replaced and the slurry cooled. The slurry was cooled to 5-10°C and granulated for 2 hours after which it was filtered and washed with 2ml/g (0.3 litres) of 2-butanone. The salt was dried overnight in vacuo at 55-60°C to yield a white crystalline solid. Yield 215g, 96.4%. Mpt 242-244°C. δ (DMSO): 1.17 (3H, t), 1.28 (3H, t), 1.35 (3H, t), 2.73 (2H, q), 2.97 (2H, q), 3.2 (3H, s), 3.58 (2H, t), 3.78 (3H, t), 3.81 (2H, t), 4.49 (2H, t) 4.51 (2H, q), 7.29-7.33 (3H, m), 7.57-7.60 (2H, m), 8.28 (1 H, d), 8.73 (1 H, d), 9.13 (1 H,s), 11.90(1 H,s).
The powder X-ray diffraction (PXRD) pattern for this salt, having Mpt 242- 244°C, was determined using a Siemens D5000 powder X-ray diffractometer fitted with a theta-theta goniometer, automatic beam divergence slits, a secondary monochromator and a scintillation counter. The specimen was rotated whilst being irradiated with copper K-alpha1 X-rays (Wavelength = 1.5046 Angstroms) filtered with a graphite monochromator (λ = 0.15405nm) with the X-ray tube operated at 40 kV/mA. The main peaks (in degrees θ) of the PXRD pattern are illustrated in Table I.
Table


The same besylate salt, as defined by the XRD pattern described in Table 1 , when made via alternative routes can have a melting point in the range of from 235-246°C (measured using a Perkin Elmer DSC7 at a heating rate of 20°C/minute).

References
1 The discovery of UK-369003, a novel PDE5 inhibitor with the potential for oral bioavailability and dose-proportional pharmacokinetics
Bioorg Med Chem 2012, 20(1): 498………….MP 161 – 162 °C
2. Hajikarimian, Y.; Yeo, S.; Ryan, R.W.; Levett, P.; Stoneley, C.; Singh, P.
Investigation into the formation of the genotoxic impurity ethyl besylate in the final step manufacturing process of UK-369,003-26, a novel PDE5 inhibitor
Org Process Res Dev 2010, 14(4): 1027
3. Bentham; Dawson; Dunn; Papadopoulos; Taylor; Mitchell; Snowden; Taylor
Organic Process Research and Development, 2004 , vol. 8, 4 PG. 674 – 679 ………….AS ENTRY B
- Bloch, W., et al.: Prostate, 33, 1 (1997)
- • Glowienke, S., et al.: Mutat. Res., 581, 23 (1997)
- • Chapple, C., et al.: Eur. Urol., 54, 563 (1997)
- • Elder, D., et al.: J. Pharm. Pharmacol., 61, 269 (1997)
PATENTS
1. WO 2010062366
2. WO 2007072156
3 WO 2007072156
4.US2002/22732 A1,
5.US2002/28799 A1,
6.
| WO1998049166A1 * | Apr 10, 1998 | Nov 5, 1998 | Mark Edward Bunnage | PYRAZOLOPYRIMIDINONES WHICH INHIBIT TYPE 5 CYCLIC GUANOSINE 3′,5′-MONOPHOSPHATE PHOSPHODIESTERASE (cGMP PDE5) FOR THE TREATMENT OF SEXUAL DYSFUNCTION |
| WO1999054333A1 * | Mar 25, 1999 | Oct 28, 1999 | Mark Edward Bunnage | Pyrazolopyrimidinone cgmp pde5 inhibitors for the treatment of sexual dysfunction |
| US4666921 * | 15 окт 1985 | 19 май 1987 | Ludwig Heumann & Co. Gmbh | Pyrazole derivatives, processes for their preparation and pharmaceutical preparations containing these compounds |
| US5808092 * | 15 окт 1997 | 15 сен 1998 | Ube Industries, Ltd. | Process for preparing-1-ethyl-5-hydroxypyrazole |
| US6015911 * | 24 мар 1998 | 18 янв 2000 | Dow Agrosciences Llc | Process for preparing 1-alkyl-4-(2-chloro-3-alkoxy-4-alkylsulfonylbenzoyl)-5-hydroxypyrazole and related compounds |
| EP0463756A1 | 7 июн 1991 | 2 янв 1992 | Pfizer Limited | Pyrazolopyrimidinone antianginal agents |
| EP0812845A1 | 4 июн 1997 | 17 дек 1997 | Pfizer Limited | Process for preparing sildenafil |
| EP0994115A2 | 11 окт 1999 | 19 апр 2000 | Pfizer Limited | Process for preparation of pyrazolo-(4,3-d)pyrimidin-7-ones and intermediates thereof |
| EP0995750A1 | 15 окт 1999 | 26 апр 2000 | Pfizer Inc. | Pyrazolopyrimidinone cGMP PDE5 inhibitors for the treatment of sexual dysfunction |
| WO1998049166A1 | 10 апр 1998 | 5 ноя 1998 | Mark Edward Bunnage | PYRAZOLOPYRIMIDINONES WHICH INHIBIT TYPE 5 CYCLIC GUANOSINE 3′,5′-MONOPHOSPHATE PHOSPHODIESTERASE (cGMP PDE5) FOR THE TREATMENT OF SEXUAL DYSFUNCTION |
| WO1999054333A1 | 25 мар 1999 | 28 окт 1999 | Mark Edward Bunnage | Pyrazolopyrimidinone cgmp pde5 inhibitors for the treatment of sexual dysfunction |
| WO2001027112A1 | 4 окт 2000 | 19 апр 2001 | Charlotte Moira Norfo Allerton | 5-(2-substituted-5-heterocyclylsulphonylpyrid-3-yl)-dihydropyrazolo[4,3-d]pyrimidin-7-ones as phosphodiesterase inhibitors |
| WO2001027113A2 | 11 окт 2000 | 19 апр 2001 | Mark Edward Bunnage | PYRAZOLO `4,3-d! PYRIMIDINE DERIVATIVES |
PDE5 inhibitors mirodenafil

sildenafil

tadalafil

udenafil 3-(l-methyl-7-oxo-3-propyl-4H-pyrazolo[5,4-e]pyrimidin-5-yl)-N- [2-(l -methylpyrrolidin-2-yl)ethyl] -4-propoxybenzenesulfonamide

vardenafil 4-[2-ethoxy-5-(4-ethylpiperazin-l-yl)sulfonyl-phenyl]-9-methyl-7- propyl- 3,5,6,8-tetrazabicyclo[4.3.0]nona-3,7,9-trien-2-one

avanafil 4-[(3-chloro-4-methoxy-phenyl)methylamino]-2-[(2S)-2- (hydroxymethyl)pyrrolidin- 1 -yl] -N-(pyrimidin-2- ylmethyl)pyrimidine-5-carboxamide

dasantafil 7-[(3-bromo-4-methoxyphenyl)methyl]-l-ethyl-8-[[(lR,2R)-2- hydroxycyclopentyl]amino]-3-(2-hydroxyethyl)purine-2,6-dione
NM 702 (Nissan Chemical Industries)

SLX 101 (Surface Logix) – Structure Not Available
UK 369003 (Pfizer) – Gisadenafil besylate


334827-98-4 (as besylate)
- UK 369003
- UK-369,003
- UK0369,003
- UNII-S6G4R7DI1C
symptoms associated with BPH

 1…………..


7…………………


1.07 (3H, t), METHYL OF -N CH2-CH3 ON PIPERAZINE RING
1.42 (3H, t), METHYL OF -CH2-CH3 ON PYRAZOLE SIDE CHAIN
1.61 (3H, t), METHYL OF -O-CH2-CH3 ON PYRIMIDINE RING
2.44 (2H, q), CH2 OF -N CH2-CH3 ON PIPERAZINE RING
2.57 (4H, m),4H OF –NCH2 ON PIPERAZINE RING BOTH SIDE OF N ATOM
3.08 (2H, q), CH2 OF –CH2-CH3 ON PYRAZOLE SIDE CHAIN
3.15 (4H, m),4H OF –NCH2 ON PIPERAZINE RING BOTH SIDE OF N ATOM CLOSE TO SO2 GP
3.32 (3H, s),METHYL OF -OCH3 ON PYRAZOLE SIDE CHAIN
3.92 (2H, q), CH2 OF NCH2-CH2-O-CH3 ON PYRAZOLE SIDECHAIN
4.48 (2H, q), CH2 OF NCH2 –CH2-O-CH3 ON PYRAZOLE SIDECHAIN
4.77 (2H, q), CH2 OF O-CH2 CH3 ON PYRIMIDINE RING
8.65 (1H, d), PYRIMIDINE AROM H …..AWAY/PARA TO C=O-NH -PYRAZOLE GP
9.06 (1H, d). PYRIMIDINE AROM H …..CLOSER/ORTHO TO C=O-NH -PYRAZOLE GP, reason this signal will shift from 8,86 delta to 9.06 after cyclization in this step ie formation of GISADENAFIL
The spectrum also has signals that correspond to a solvate with ethyl acetate.
Orphan Drugs: Global Regulatory Events

This Blog Post discusses recent global non-US regulatory events for orphan drugs.
I – Europe
At a January meeting, the European Medicines Agency’s (EMA) Committee for Medicinal Products for Human Use (CHMP) gives the following opinions for three orphan drugs :
• Positive recommendation for Bayer’s Adempas (Riociguat) for the treatment of Chronic Thromboembolic Pulmonary Hypertension (CTEPH) and Pulmonary Arterial Hypertension (PAH)
• Negative opinion for AB Science’s Masiviera (Masitinib) which is intended for the treatment of non resectable locally advanced or metastatic pancreatic cancer
• Negative opinion for PTC Therapeutics’ Translarna (Ataluren) which is intended for the treatment of Duchenne Muscular Dystrophy.
Also, EMA’s Committee for Orphan Medicinal Products (COMP) issues 15 positive opinions for ODD at their January meeting (Reference Blog Post). These ODDs are to be presented to the European Commission (EC) for final approval. If the EC approves these ODDs, the drugs receive ODD in the…
View original post 230 more words
UDENAFIL …The Eastern Viagra (like)

UDENAFIL
An oral phosphodiesterase 5 inhibitor used for the treatment of erectile dysfunction.
268203-93-6 CAS NO
LAUNCHED 2005 MEZZION DA-8159 ME-3113 Udzire Zydena MEZZION …INNOVATOR
POWERPOINT PRESENTATION BY INNOVATOR.. CLICK HERE
| Synonyms: | Zydena;Udenafi;Da-8159;Da 8159;Udenafil;Udenafil(DA 8159,Zydena);5-(2-Propyloxy-5-(1-methyl-2-pyrollidinylethylamidosulfonyl)phenyl)-1-methyl-3-propyl-1,6-dihydro-7H-pyrazolo(4,3-D)pyrimidine-7-one;5-[2-propyloxy-5-[2-(1-Methyl-2-pyrrolidinyl)ethylaMinosulfonyl]phenyl]-1-Methyl-3-propyl-1,6-dihydro-7H-pyrazolo[4,3-d]pyriMidine-7-one;5-[2-propyloxy-5-(2-(1-Methylpyrrolidin-2-yl)ethylaMinosulphonyl)phenyl]-1-Methyl-3-propyl-6,7-dihydro-1H-pyrazolo(4,3-d)pyriMidin-7-one;3-(6,7-Dihydro-1-methyl-7-oxo-3-propyl-1H-pyrazolo[4,3-d]pyrimidin-5-yl)-N-[2-(1-methyl-2-pyrrolidinyl)ethyl]-4-propoxybenzenesulfonamide |
| Molecular Formula: | C25H36N6O3S2 |
| Formula Weight: | 516.66 |
3-(1-methyl-7-oxo-3-propyl-4,7-dihydro-1H-pyrazolo[4,3-d]pyrimidin-5-yl)-N-[2-(1-methylpyrrolidin-2-yl)ethyl]-4-propoxybenzenesulfonamide
(5- [2-propyloxy-5- (1- methyl-2-pyrolidinylethylamidosulfonyl) phenyl] -1-methyl- propyl-1, β-dihydro-7H-pyrazolo (4 , 3-d) pyrimidin-7-one)
A pyrazolo-pyrimidinone similar to sildenafil; phosphodiesterase type 5 inhibitor. Udenafil is a new phosphodiesterase type 5 (PDE5) inhibitor used to treat erectile dysfunction (ED). It has been approved in South Korea and will be marketed under the brand name Zydena.
It is not yet approved for use in the U.S., E.U., or Canada. Udenafil (Zydena®) is also a potent and selective PDE5i developed by Dong-A Pharmaceutical Company in Korea (Kim et al., 2008; Han et al., 2010).
It has not yet been approved by FDA or the European Medicines Agency (EMEA) and was only approved by the Korean Food and Drug Administration (KFDA), being currently used in Korea and Russia (Alwaal et al., 2011; Cho et al., 2012).
- DA 8159
- DA-8159
- Udenafil
- UNII-L5IB4XLY36
- Zydena
Udenafil is a drug used in urology to treat erectile dysfunction. It belongs to a class of drugs called PDE5 inhibitor, which many other erectile dysfunction drugs such as sildenafil, tadalafil, and vardenafil also belong to. It was developed by Dong-A Pharmaceutical Co., Ltd. and is marketed under the trade name Zydena™.[2] With a T max of 1.0-1.5 h and a T 1/2 of 11-13 h (a relatively rapid onset and a long duration of action), both on-demand and once-daily use of udenafil have been reported.[3] Typical doses are 100 and 200 mg. It is not approved for use in the United States by theU.S. Food and Drug Administration. Udenafil (DA-8159), a pyrazolopyramidinone derivative that acts as a phosphodiesterase 5 (PDE5) inhibitor, was launched by Dong-A Pharmtech (currently Mezzion Pharma) in late 2005 in Korea for the oral treatment of erectile dysfunction (ED). The company is currently conducting phase III clinical trials in the U.S. for this indication.
Dong-A Pharmatech is conducting phase III clinical trials for the treatment of patients with portal hypertension resulting from liver disease and for the treatment of benign prostatic hyperplasia (BPH). Phase II/III clinical studies at Dong-A Pharmatech for the treatment of secondary Raynaud phenomenon have been completed. Meiji Seika Pharma is developing the compound in phase I clinical trials for the treatment of BPH in Japan.
Phosphodiesterases regulate the tissue concentration of cyclic guanosine monophosphate (cGMP), which in turn triggers smooth muscle relaxation, allowing blood to flow into the penis and resulting in erection. PDE5 is the most abundant phosphodiesterase in the human corpus cavernosum, and as such its inhibition by DA-8159 enhances erectile function by increasing the concentration of cGMP. Results from phase I studies indicate that udenafil has a unique pharmacokinetic profile with a relatively rapid onset and sufficiently long duration to make it effective for up to 24 hours. In 2009, the compound was licensed to Warner Chilcott (acquired by Actavis in 2013) by Dong-A Pharmatech for development and marketing in the U.S. for the oral treatment of erectile dysfunction.
In 2011, udenafil was licensed to Meiji Seika Pharma by Dong-A ST in Japan for the treatment of benign prostatic hyperplasia. Udenafil is a potent novel phosphodiesterase-5 inhibitor approved for use in Korea. Udenafil has unique properties, with a T max of 1.0–1.5 h and a T 1/2 of 11–13 h (a relatively rapid onset and a long duration of action). Therefore, both on-demand and once-daily use of udenafil have been reported. Udenafil’s efficacy and tolerability have been evaluated in several studies, and recent and continuing studies have demonstrated udenafil’s promise in both dosing regimens. Presently, tadalafil is the only FDA-approved drug for daily dosing, but udenafil can be used as a once-daily dose for erectile dysfunction patients who cannot tolerate tadalafil due to phosphodiesterase subtype selectivity. Udenafil as an on-demand or once-daily dose is effective and tolerable, but more studies are needed in patients of other ethnicities and with comorbid conditions such as diabetes mellitus, hypertension, and benign prostate hyperplasia.
Erectile dysfunction (ED) is defined as the inability to achieve and maintain a sufficient erection to permit satisfactory intercourse [Montorsi et al. 2010]. Numerous strategies have been used to overcome ED. Therapies for ED include intracavernosal injection, vacuum erection devices, intraurethral suppositories, penile prosthesis surgery and oral phosphodiesterase-5 (PDE5) inhibitors [Dinsmore and Evans, 1999]. Oral PDE5-inhibitor medications have revolutionized the treatment of ED. Men prefer oral medications as the first-line therapeutic option in the absence of a specific contraindication to their use [Ding et al. 2012].
There are currently four PDE5 inhibitors (sildenafil, vardenafil, tadalafil, and avanafil) approved worldwide for the treatment of male erectile dysfunction, with two other agents (udenafil and mirodenafil) currently approved only in Korea [Bell and Palmer, 2011]. The choice of PDE5 inhibitor for each patient should be determined after physician and patient discuss the characteristics of different drugs and the individual patient’s sexual habits, preferences, and expectations [Hatzimouratidis et al. 2010]. There are two types of treatment usage of PDE5 inhibitors according to their pharmacological characteristics. On-demand treatment of ED with PDE5 inhibitors allows the patient to have intercourse within 1 hour, but can remove spontaneity from sexual activity and be burdensome to patients and their partners [Hanson-Divers et al. 1998]. Once-daily dosing of a PDE5 inhibitor is an alternative for couples that prefer spontaneous sexual activities.
A new oral selective PDE5 inhibitor, udenafil (Zydena, Dong-A, Seoul, Korea), has recently been developed for the treatment of ED. Udenafil is a novel pyrazolopyrimidinone compound developed by Dong-A Pharmaceutical Co., Ltd (Seoul, Korea) for the treatment of ED which has the same mechanism of action as sildenafil [Kim et al. 2008]. Udenafil is rapidly absorbed, reaching peak plasma concentrations at 0.8–1.3 h, then declining monoexponentially with a terminal half-life (T 1/2) between 7.3 and 12.1 hours, giving it the unique pharmacokinetics of both relatively rapid onset and long duration [Salem et al. 2006]. Thus, both on-demand treatment and once-daily dosing have been reported in the literature. The purpose of this review is to evaluate the efficacy and tolerability of udenafil for patients with ED according to the currently available literature.
Udenafil” refers to the chemical compound, 3-(1-methyl-7-oxo-3-propyl-4,7-dihydro-1H-pyrazolo[4,3-d]pyrimidin-5-yl)-N-[2-(1-methylpyrrolidin-2-yl)ethyl]-4-propoxybenzenesulfonamide and has the following chemical formula:

More information about udenafil can be found at Kouvelas D. et al., (2009) Curr Pharm Des, 15(30):3464-75. Udenafil is marketed under the trade name Zydena® but not approved for use in the United States. TRADE NAME IN INDIA – UDEZIRE Erectile dysfunction (ED) is an inability to achieve or sustain an erection suitablefor sexual intercourse.
Sexologists say at least 10% men in India may have to use these drugs at some point. Studies have shown that 40% of men up to the age of 40 years have ED andit goesup 70% by 60 years. The commonly prescribed drugs for the disorder in India are sildenafil(Viagra) and tadalafil,which belong to a category called phosphodiesterasetype5 drugs.
Now, Zydus, a pharmaceutical company, has got exclusive permission to sell udenafil. It’s not always that the release of a drug is celebrated by many, particularly men. A drug that was released in India last week is the recent in the list of drugs that has a cure for erectile dysfunction. The manufacturers say udenafil, which will be marketed under the brand name Udezire, will be long-acting, but with minimal side effects. Erectile dysfunction (ED) is an inability to achieve or sustain an erection suitable for sexual intercourse. Sexologists say at least 10% men in India may have to use these drugs at some point. Studies have shown that 40% of men up to the age of 40 years have ED and it goes up 70% by 60 years

UDENAFIL
……………………
INTRODUCTION
Udenafil (Zydena®) is a therapeutic agent hypothesized to improve erectile function endpoints through interaction with the phosphodiesterase type 5 (PDE5) enzyme. As such, udenafil belongs to the class of such agents that includes tadalafil (Clalis®), sildenafil (Viagra®), and vardenafil (Levitra®). These agents are purported to promote erectile response through inhibition of PDE5, the predominant PDE within the penis, which leads to higher intracellular levels of cyclic guanylate cyclase (cGMP). cGMP is a second messenger for the smooth-muscle relaxing effects of nitric oxide within the penis. The various agents differ in pharmacology primarily based on 1) onset and duration of action and 2) selectivity profiles vs. other PDEs. All three marketed agents have proven remarkably safe. These agents should not be taken by patients with unstable cardiovascular disease. Udenafil has been shown to exhibit greater selectivity against the known PDE homologues, than other PDE5 inhibitors. Udenafil is comparable to tadalafil in many respects, such as duration of action and high selectivity for PDE6, but udenafil has greater selectivity for PDE11 than tadalafil.
Tadalafil, with a half life of 17.5 hours, has a much longer duration of action and improved exercise tolerance than either sildenfail or vardenafil, which have half lives of 4-5 hours. Consequently, tadalafil is associated with less planning or pressure to have sexual intercourse after dosing. Dissociation of the sexual activity from the time of dosing is associated with higher rates of patient and partner satisfaction. In prospective, randomized crossover clinical studies, patients preferred tadalafil over sildenafil by margins ranging from 7:3 to 9:1. Sildenafil and vardenafil both modulate PDE6 at higher rate than tadalafil. PDE6 modulation has been associated with chromatopsia. The side effects of chromatopsia, such as sensitivity to light and blurred vision, are therefore higher in patients taking sildenafil or vardenafil, about 2-3%, than patients taking tadalafil, about <0.1%. Tadalafil is less selective than sildenafil and vardenafil for PDE5 and for PDE11a. Activity at PDE11a is suspected to have a causal relationship with myalgia and testicular toxicity. The selectivity profile for udenafil is similar to sildenafil, which should impart greater safety for this agent.

The benefits and shortcomings of these drugs have been reviewed. Some of these shortcomings can be traced to metabolism-related phenomena. Udenafil is converted in vivo by oxidative and conjugative degradation to multiple metabolites. Phase I metabolism leads to demethylation of the pyrazole, hydroxylation of the pyrazole propyl group, and dealkylation alpha to the sulfonamide nitrogen to afford an active metabolite. Because udenafil is metabolized primarily by cytochrome P450 subtype 3A4 (CYP3A4), exposure to udenafil can influence polypharmacy. For example, CYP3A4 inhibitors such as HIV protease inhibitors, azole antifungals, and erythromycin can lead to higher than otherwise expected blood levels of udenafil. Conversely, co-administration of CYP3A4 inducers such as rifampin can decrease the otherwise expected blood levels of udenafil. Thus, the polypharmacy of udenafil is necessarily complex and has potential for adverse events. In addition, there may be increased inter-patient variability in response to polypharmacy.
Analogs of udenafil as described herein have the potential to alleviate the problems associated with the commercially available PDE5 inhibitors while maintaining or improving efficacy. It is believed that the reduction in CYP3A4 clearance of udenafil analogs will be expected to increase the proportion of clearance via mechanisms less susceptible to polypharmaceutical complications. In addition, analogs of udenafil having an attenuated rate of oxidative metabolism will have an increased half-life, further augmenting their advantages vs. tadalafil, sildenafil and vardenafil. Potentially, a single dose of an udalafil analog, described herein, having an increased half-life may provide therapeutic coverage for an entire weekend or beyond while increasing safety parameters by reducing the likelihood of drug-drug interactions and by increasing safety as a result of the increased selectivity.

The compounds of formula 1 may contain asymmetric centers and thus they can exist as enantiomers. The present invention includes both mixtures and separate individual isomers . Male erectile dysfunction is one of the most common sexual dysfunctions in men. Although erectile dysfunction can be primarily psychogenic in origin, it often accompanies chronic illnesses, such as diabetes mellitus, heart disease, hypertension, and a variety of neurological diseases. Its prevalence is strongly related to age, with a estimated prevalence of 2% at age 40 years rising to 25-30% by age of 65. Although no data are available on the prevalence of erectile dysfunction in men aged over 75, it is probably over 50%. Various treatment options for erectile dysfunction are available, such as counseling, hormonal therapy, self-injection or transurethral application of vasodilator agents, vacuum devices, prosthesis implantation, and venous/arterial surgery. However, these therapeutic options have several limitations such as side effects, high-cost and low efficacy.
Therefore it has called for research efforts to develop new, high effective and simple to use treatment methods, potentially oral medication. Recently, sildenafil has been developed as a therapeutic agent for male erectile dysfunction by oral administration. Sildenafil is the first in a new class of drugs known as inhibiting phosphodiesterase-5 enzyme distributed specifically in corpus cavernosal tissues and induces relaxation of the corpus cavernosal smooth muscle cells, so that blood flow to the penis is enhanced, leading to an erection.
Sildenafil has shown a response rate of around 80% in men with erectile dysfunction of organic cause. On the other hand, USP 3,939,161 discloses that 1 , 3 -dimethyl -lH-pyrazolopyrimidinone derivatives exhibit anticonvulsant and sedative activiity, and also exhibit anti-inflammatory activity and gastric antisecretory activity; EP 201,188 discloses that 5-substituted pyrazolopyrimidinone derivatives have effects of antagonizing adenosine receptor and of inhibiting phosphodiesterase enzymes and can be used for the treatment of cardiovascular disorders such as heart failure or cardiac insufficiency; EP 463,756, EP 526,004, WO 93/6,104 and WO 93/7,149 disclose that pyrazolopyrimidinone derivatives which inhibit c-GMP phosphodiesterase more selectively than c-AMP phosphodiesterase have efficacy on cardiovascular disorders such as angina pectoris, hypertension, heart failure, atherosclerosis, chronic asthma, etc.; and WO 94/28,902, WO 96/16,644, WO 94/16,657 and WO 98/49,166 disclose that the known inhibitors of c-GMP phosphodiesterase including the pyrazolopyrimidinone derivatives of the above mentioned patents can be used for the treatment of male erectile dysfunction Since sildenafil has been developed, various compounds for inhibiting phosphodiesterase-5 have been reported.
Among them, pyrazolopyrimidinone compounds of formula 1 (KR Pat. No. 99-49384) were reported having better potency than that of sildenafil, based on the mechanism of inhibiting phosphodiesterase-5 and having better selectivity over phosphodiesterase-6 distributed in retina and phosphodiesterase-3 distributed in heart to reduce the side effects. Further, the pyrazolopyrimidinone compounds of formula 1 were said to be improved the solubility and the metabolism in the liver, which are very important factor affecting the rate of the absorption when administered orally.
The KR patent No. 99-49384 also disclosed a process for preparing the pyrazolopyrimidinone compounds of formula , comprising the steps of: a) reacting chlorosulfonated alkoxy bonzoic acid with a primary amine to obtain sulfonamide-substituted benzoic acid; b) reacting the obtained sulfonamide-substituted benzoic acid with pyrazolamine in the presence of activating reagent of carboxylic group or coupling agent of carboxylic group with amine group to obtain corresponding amide compound; and, c) performing an intramolecular cyclization of the obtained amide compound to obtain the pyrazolopyrimidinone compound of formula 1. This reaction is represented in scheme 1 Scheme 1

…………………..
SYNTHESIS
The present invention provides an agent comprising a pyrazolopyrimidinone compound (5- [2-propyloxy-5- (1- methyl-2-pyrolidinylethylamidosulfonyl) phenyl] -1-methyl- propyl-1, 6-dihydro-7H-pyrazolo (4, 3-d) pyrimidin-7-one) expressed as formula 1 as an effective ingredient for preventing and treating benign prostatic hyperplasia (BPH) . Formula 1

The pyrazolopyrimidinone compound represented as formula 1 is one of the PDE-5 inhibitors and has characteristics in that it has a strong inhibitive activity and an excellent selectivity for PDE-5; it is readily absorbed as its solubility is improved; it has a good bioavailability and a large volume of distribution; and it has an in vivo half-life longer three times than sildenafil or vardenafil, a drug of the same mechanism. Physicochemical properties of the pyrazolopyrimidinone compound of formula 1 are as follows: it is hardly dissolved in water; however, it is readily dissolved in acetic acid, methanol, chloroform and the like; and it is a white or pale yellow powder, not a hydrate or a solvate, having a melting point of 158 to 161 “Q and having pKal and pKa2 of about 6.5 and 12.5, respectively. The pyrazolopyrimidinone compound represented as formula 1 is prepared via a synthetic process consisting of roughly three steps. The inventors of the present invention have disclosed a method for preparing the same in WO2000/027847 (Corresponding Korean Patent No.0353014), which will now be described roughly as follows. First, in the first step, 4- [2-propyloxy-5- (chlorosulfonyl) benzamido] -l-methyl-3-propyl-5-carbamoyl pyrazole is prepared.
For such preparation, a specified amount of 4- [2-propyloxybenzamido] -l-methyl-3-propyl-5- carbamoyl pyrazole is added to a specified amount of chlorosulfonic acid cooled to 0 °Q then, the resultant mixture is stirred, filtered, washed and dried to obtain 4- [2-propyloxy-5- (chlorosulfonyl) benzamido] -l-methyl-3- propyl-5-carbomoyl pyrazole. In the second step, from the pyrazole compound prepared in the first step, 4- [2-propyloxy-5- ( l-methyl-2- pyrolidinylethylamidosulfonyl) benzamido] -l-methyl-3- propyl-5-carbomoyl pyrazole is prepared. For such preparation, a specified amount of 2- (2-aminoethyl) -1- methyl pyrolidine is added in dichloromethane solution of the specified amount of 4- [2-propyloxy-5- (chlorosulfonyl) benzamido] -l-methyl-3-propyl-5-carbamoyl pyrazole prepared in the first step to be stirred. Then, the reactant solution is diluted with dichloromethane. The organic layer is washed, dried, concentrated and filtered to obtain 4- [2-propyloxy-5- (l-methyl-2- pyrolidinylethylamidosulfonyl) benzamido] -l-methyl-3- propyl-5-carbomoyl pyrazole is obtained.
Last, in the third step, the pyrazolopyrimidinone compound of the present invention (5- [2-propyloxy-5- (1- methyl-2-pyrolidinylethylamidosulfonyl) phenyl] -1-methyl- propyl-1, β-dihydro-7H-pyrazolo (4 , 3-d) pyrimidin-7-one) is prepared from the compound obtained in the second step. For such preparation, the specified amount of pyrazole compound prepared in the second step is dissolved in t- butanol . A specified amount of potassium t-butoxide is added in the resultant solution and, then, reflux-stirred for a predetermined time. After the resultant solution is cooled, diluted, washed and dried, distillation under reduced pressure, solvolysis and silica gel column chromatography are carried out, thus obtaining a specified amount of pure pyrazolopyrimidinone compound of the present invention
. …………………………..
SYNTHESIS WO2000027848A1
REACTION SCHEME 2

The process for preparation according to the present invention comprises the steps of : 1) reacting the chlorosulfonated compound of formula ( 2 ) and primary amine (3_) under the condition of suitable temperature and suitable solvent to give sulfonamide (4.) (step 1) ; 2) reacting the carboxylic acid (4.) prepared in step 1 and pyrazoleamine (5) to give an amide (6.) by the known method preparing amide from carboxylic acid and amine (step 2) ; and 3) cyclizing the amide (6.) prepared in step 2 to give the desired compound of formula 1 by the known cyclization method used for preparation of pyrimidinone (step 3) .
In step 1, a little excess of 2 equivalents of amine may be used, or a little excess of 1 equivalent of amine and 1 equivalent of acid scavenger such as tertiary amine are may be used together. The reaction temperature is preferred below 20 °C. The known method preparing amide from carboxylic acid and amine in step 2 is the process, for example, in which carboxyl group is transformed into activated acid chloride or acid anhydride by using thionyl chloride, pivaloyl chloride, trichlorobenzoyl chloride, carbonyldiimidazole, diphenylphosphinic chloride, etc. and followed by reacting with amine group, or the process using coupling agents such as DCC (1,3-dicyclo hexylcarbodiimide) or EEDQ (N-ethoxycarbonyl -2 -ethoxy- 1, 3-dihydroquinoline) .
The cyclization process in step 3 may be carried out in the presence of a suitable base and a suitable solvent. Preferred bases which are employed in step 3 are metal alkoxides; metal salts of ammonia; amine; hydrides of alkali metal or alkaline earth metal; hydroxides; carbonates; bicarbonates ; and bicyclic amidines such as DBU (1 , 8-diazabicyclo [5.4.0] undec -7-ene) and DBΝ (1 , 5-diazabicyclo [4.3.0] non-5-ene) . Preferred solvents which are employed in step 3 are alcohols such as methanol, ethanol, isopropanol, t-butanol, etc.; ethers such as tetrahydrofuran, dimethoxyethane, dioxane, etc.; aromatic – hydrocarbons such as benzene, toluene, xylene, chlorobenzene, etc.; acetonitrile; dimethylsulfoxide; dimethylformamide; N-methylpyrrolidin-2 -one ; and pyridine.
SEE ENTRY no 68
5- [2-propyloxy-5- ( 1-methyl-2-pyrrolidinylethyl amidosulfonyl) phenyl] -l-methyl-3 -propyl-1 , 6-dihydro-7 H-pyrazolo (4 , 3-d) yrimidin-7-one (compound of example68) 
ACCORDING TO ME ENTRY IS 68 ANY ERROR, amcrasto@gmail.com
Synthesis WO2001098304A1
The present invention relates to a process for preparing pyrazolopyrimidinone derivatives of formula 1 and pharmaceutically acceptable salts thereof which have an efficacy on impotence, comprising the steps of chlorosulfonation of pyrazolamide compounds of formula 2, followed by amination with a primary amine and intramolecular cyclization. Formula 1

Formula 2

The compounds of formula 1 may exist in tautomeric equilibrium as shown below.

The compounds of formula 1 may also contain asymmetric centers and thus they can exist as enantiomers. The present invention includes both racemic mixture and separate individual enantiomers. Scheme 2

……………………………….
SYNTHESIS WO2010013925A2
INTERMEDIATES
4-[2-propyloxy benzamido]-l-methyl-3-propyl-5-carbamoyl pyrazole
CHLOROSULPHONIC ACID
4-[2-propyloxy-5-(chlorosulfonyl)benzamido]-l-methyl-3-propyl-5-carbamoyl pyrazole
2-(2-aminoethyl)-l-methylpyrrolidine 4-[2-propyloxy-5-(l-methyl-2-pyrrolidinylethyl amido- sulfonyl)benzamido]-l-methyl-3-propyl-5-carbamoyl pyrazole
potassium t-butoxide
3, 5-[2-propyloxy-5-(l-methyl-2-pyrrolidinylethyl amido- sulfonyl)phenyl]-l-methyl-3-propyl-l,6-dihydro-7H-pyrazolo(4,3-d)pyrimidin-7-one UDENAFIL
The present invention provides a pharmacological compound containing 5- [2-propyloxy-5-( 1 -methyl-2-pyrolidinylethylamidosulphonyl)phenyl] – 1 -methyl-prop yl- 1 ,6-dihydro-7H-pyrazolo(4,3-d)pyrimidin-7-one, a pyrazolopyrimidinone compound, represented by the following Chemical Formula 1 or pharmaceutically acceptable salts thereof, as an active ingredient for prevention and treatment of respiratory diseases. [14] [Chemical Formula 1]

………………
SYNTHESIS
EXAMPLE 2 3-(1-Methyl-7-oxo-3-propyl-6,7-dihydro-1H-pyrazolo[4,3-d]pyrimidin-5-yl)-N-[2-(1-methyl-pyrrolidin-2-yl)-ethyl]-4-propoxy-benzenesulfonamide

Step 1

2,4-Dioxo-heptanoic acid methyl ester: Sodium (25.3 g, 1.1 mol) was proportionally added to ethanol (350 mL) at ambient temperature with vigorous stirring, and the solution was cooled to 0° C. Pentan-2-one (86 g, 1.0 mol) and diethyl oxalate (146 g, 1.0 mol) were added sequentially at 0° C., and stirring was continued for 1 hour at 0° C., and overnight at ambient temperature. The solvent was removed under reduced pressure, diethyl ether (200 mL) and cold dilute hydrochloric acid (500 mL) were added. Following standard extractive work up, the solvent was evaporated under reduced pressure to yield the title compound (141 g, 76%). 1H-NMR (300 MHz, CDCl3) δ 14.51 (broad s, 1H), 6.37 (s, 1H), 4.35 (q, 2H, J=6.6 Hz), 2.47 (t, 2H, J=7.2 Hz), 1.76-1.66 (m, 2H), 1.38 (t, 3H, J=7.2 Hz), 0.97 (t, 3H, J=7.5 Hz); GC-MS: 186 (M)+, 113 (M-73)+
Step 2

5-Propyl-2H-pyrazole-3-carboxylic acid ethyl ester: Hydrazine hydrate (41.4 g, 827 mmol) was slowly added to a solution of 2,4-dioxo-heptanoic acid methyl ester (140 g, 753 mmol) in 280 mL of acetic acid at 0° C. The mixture was heated to reflux for 8 hours and cooled. The solvent was removed under reduced pressure; the residue was diluted with diethyl ether (300 mL). Following standard extractive work up, the solvent was evaporated under reduced pressure to yield the title compound as a white solid (131 g, 96%). 1H NMR (300 MHz, CDCl3) δ 9.27 (broad s, 1H), 6.61 (s, 1H), 4.37 (q, 2H, J=7.2 Hz), 2.68 (t, 2H, J=7.5 Hz), 1.75-1.62 (m, 2H), 1.37 (t, 3H, J=6.6 Hz), 0.96 (t, 3H, J=7.2 Hz); LC-MS: m/z=183 (MH)+;
Step 3

2-Methyl-5-propyl-2H-pyrazole-3-carboxylic acid ethyl ester: A mixture of 5-propyl-2H-pyrazole-3-carboxylic acid ethyl ester (32.8 g, 180 mmol) and dimethyl sulfate (24.9 g, 198 mmol) was heated at 90° C. for 3 hours. The reaction was cooled and diluted with dichloromethane (200 mL). Following standard extractive work up, the solvent was evaporated under reduced pressure to yield a crude residue which was purified by flash chromatography on silica gel to give the title compound as a colorless oil (23 g, 65%). 1H NMR (300 MHz, CDCl3) δ 6.59 (s, 1H), 4.37 (q, 2H, J=7.2 Hz), 2.58 (t, 2H, J=7.2 Hz), 1.76-1.64 (m, 2H), 1.40 (t, 3H, J=6.6 Hz), 1.01 (t, 3H, J=7.2 Hz), 4.40 (q, 2H), 3.89 (s, 3H), 2.59 (t, 2H), 1.69 (2H), 1.37 (t, 3H), 1.01 (t, 3H); LC-MS: m/z=197 (MH)+.
Step 4

2-Methyl-5-propyl-2H-pyrazole-3-carboxylic acid: 2-methyl-5-propyl-2H-pyrazole-3-carboxylic acid ethyl ester (29.4 g, 150 mmol) was suspended in 6N sodium hydroxide (120 mL, 720 mmol) and heated to 80° C. for 2 hours, cooled, diluted with water (100 mL) and acidified with 5N hydrochloric acid (200 mL) to give a precipitate which was filtered off and dried to give the title compound as a white solid (24.2 g, 96%). 1H NMR (300 MHz, CDCl3) δ 6.76 (s, 1H), 4.17 (s, 3H), 2.63 (t, 2H, J=7.2 Hz), 1.70-1.68 (m, 2H), 0.98 (t, 3H, J=7.2 Hz); LC-MS: m/z=169 (M+H)+;
Step 5

2-Methyl-4-nitro-5-propyl-2H-pyrazole-3-carboxylic acid: A solution of 2-methyl-5-propyl-2H-pyrazole-3-carboxylic acid (22 g, 131 mmol) in concentrated sulfuric acid (98%, 85 mL) was heated to 50° C. and treated with a mixture of fuming nitric acid (95%, 7.7 mL) and concentrated sulfuric acid (98%, 18 mL), while keeping the reaction temperature between 50 and 55° C. The reaction mixture was kept for 8 hours at 50° C., cooled to ambient temperature, and slowly added to cold water (600 mL, 4° C.), keeping the temperature below 25° C. The precipitate was collected by filtration, and dried below 80° C. to give the title compound as a white solid (25 g, 90%). 1H NMR (300 MHz, CDCl3) δ 4.25 (s, 3H), 2.92 (t, 2H, J=7.5 Hz), 1.77-1.70 (m, 2H), 1.03 (t, 3H, J=7.2 Hz); LC-MS: m/z=214 (M+H)+
Step 6

2-Methyl-4-nitro-5-propyl-2H-pyrazole-3-carboxamide: To a suspension of 2-methyl-4-nitro-5-propyl-2H-pyrazole-3-carboxylic acid (17.0 g, 79.8 mmol) in dry toluene (85 mL) was added a catalytic quantity of dimethylformamide (0.6 mL). The mixture was heated to 50° C. and thionyl chloride (17.1 g, 143.7 mmol) was added over 30 minutes. The reaction was stirred and heated at 55-60° C. for 6 hours. The solvent was removed, dry toluene (80 mL) was added and the mixture was cooled to 20° C. and cold (5° C.) concentrated ammonium hydroxide (100 mL) was added. The precipitate was filtered, washed with water and dried to give the title compound as an off-white solid (14.8 g, 87%). LC-MS: m/z=213 (M+H)+, 235 (M+Na)+.
Step 7

4-Amino-2-methyl-5-propyl-2H-pyrazole-3-carboxamide: To a suspension of 2-methyl-4-nitro-5-propyl-2H-pyrazole-3-carboxamide (14.7 g, 69.3 mmol) in ethyl acetate (130 mL), was added 10% palladium on carbon (3.3 g). The mixture was reacted at 50° C. and 4 atm hydrogen pressure overnight. The reaction mixture was cooled, and the catalyst was filtered off and washed with ethyl acetate and dried over anhydrous magnesium sulfate, filtered and concentrated under reduced pressure to give the title compound (13.8 g, 98%). 1H NMR (300 MHz, CDCl3) δ 4.12 (s, 3H), 2.84 (s, 2H), 2.55 (t, 2H, J=7.2 Hz), 1.71-1.61 (m, 2H), 0.99 (t, 3H, J=7.2 Hz); LC-MS: m/z=183 (MH)+
Step 8

2-Methyl-4-(2-propoxybenzoylamino)-5-propyl-2H-pyrazole-3-carboxamide: A solution of 2-propoxybenzoic acid (13.7 g, 76.1 mmol) and thionyl chloride (36.2 g, 304.4 mmol) in dry dichloromethane (80 mL) was heated for 3 hours at reflux. The solvent and excess thionyl chloride were distilled off under reduced pressure. The residue was taken up in dry dichloromethane (60 mL) and reacted with a solution of 4-amino-2-methyl-5-propyl-2H-pyrazole-3-carboxamide (12.6 g, 69.2 mmol), dry triethylamine (7 g, 69.2 mmol) and 4-(N,N-dimethylamino)pyridine (84.5 mg, 0.7 mmol) in dry dichloromethane (200 mL) at 0° C. Stirring was maintained for 1 hour, and the reaction mixture was successively washed with water (150 mL), saturated aqueous sodium carbonate solution (200 mL) and saturated brine (200 mL). The organic layer was dried over anhydrous magnesium sulfate and filtered. The filtrate was concentrated to about 60 mL, and then hexane (150 mL) was added to give precipitate product as a white solid (22 g, 92%). 1H NMR (300 MHz, CDCl3) δ 9.47 (s, 1H), 8.28 (d, 1H, J=7.8 Hz), 7.87 (br.s, 1H), 7.57-7.52 (m, 1H), 7.16-7.05 (m, 2H), 5.53 (s, 1H), 4.20 (t, 2H, J=6.6 Hz), 4.09 (s, 3H), 2.54 (t, 2H, J=7.5 Hz), 1.97-1.85 (m, 2H), 1.69-1.26 (m, 2H), 1.07 (t, 3H, J=7.2 Hz), 0.95 (t, 3H, J=7.5 Hz). LC-MS: m/z=345 (M+H)+
Step 9

3-(5-Carbamoyl-1-methyl-3-propyl-1H-pyrazol-4-ylcarbamoyl)-4-propxy-benzenesulfonyl chloride: 2-Methyl-4-(2-propoxybenzoylamino)-5-propyl-2H-pyrazole-3-carboxamide (20 g, 58.1 mmol) was added to chlorosulfonic acid (81.3 g, 698 mmol) at 0° C. and the reaction was warmed to ambient temperature and stirred for 2 hours. The reaction mixture was poured into ice water (800 g) and mechanically stirred for 1 hour to give a white solid, which was filtered and washed with water. Following standard extractive work up, the solvent was evaporated under reduced pressure to yield the title compound (8 g, 31%). 1H NMR (300 MHz, CDCl3) δ 9.19 (s, 1H), 8.97 (s, 1H), 8.19 (t, 1H, J=8.9 Hz), 7.56 (br. s, 1H), 4.35 (t, 2H, J=6.6 Hz), 4.07 (s, 3H), 2.53 (t, 2H, J=7.5 Hz), 2.06-1.94 (m, 2H), 1.78-1.60 (m, 2H), 1.18 (t, 3H, J=7.5 Hz), 0.95 (t, 3H, J=7.2 Hz); LC-MS: m/z=443.1 (M+H)+
Step 10

2-Methyl-4-{5-[2-(1-methyl-pyrrolidin-2-yl)-ethylsulfamoyl]-2-propoxy-benzoylamino}-5-propyl-2H-pyrazole-3-carboxamide: To a solution of 3-(5-carbamoyl-1-methyl-3-propyl-1H-pyrazol-4-ylcarbamoyl)-4-propoxy-benzenesulfonyl chloride (2.12 g, 4.8 mmol) and dry triethylamine (0.5 g, 4.8 mmol) in dichloromethane (20 mL), was added 2-(2-aminoethyl)-1-methylpyrrolidine (0.6 g, 4.8 mmol) at 0° C. The reaction was warmed to ambient temperature, stirred for 1 hour at ambient temperature, and diluted with dichloromethane (40 mL). Following standard extractive work up, the solvent was evaporated under reduced pressure to yield the title compound (2.2 g) which was used directly in the next step. LC-MS: m/z=535 (M+H)+
Step 11

3-(1-Methyl-7-oxo-3-propyl-6,7-dihydro-1H-pyrazolo[4,3-d]pyrimidin-5-yl)-N-[2-(1-methyl-pyrrolidin-2-yl)-ethyl]-4-propoxy-benzenesulfonamide: Potassium tert-butoxide (0.9 g, 8.0 mmol) was added to a solution of crude 2-methyl-4-{5-[2-(1-methyl-pyrrolidin-2-yl)-ethylsulfamoyl]-2-propoxy-benzoylamino}-5-propyl-2H-pyrazole-3-carboxamide (2.14 g, 4.0 mmol) in dry tert-butanol (50 mL), and the mixture was heated to reflux for 8 hours. The reaction mixture was cooled to ambient temperature and diluted with ethyl acetate (300 mL). Following standard extractive work up, the solvent was evaporated under reduced pressure to yield a crude residue which was purified by flash chromatography to give the title compound (1.1 g, 53%).
1H NMR (300 MHz, CDCl3) δ 10.90 (broad s, 1H), 8.93 (s, 1H), 7.96 (d, 1H, J=8.7 Hz), 7.15 (d, 1H, J=8.7 Hz), 4.28-4.24 (m, 3H), 4.24 (s, 2H), 3.13 (t, 3H, J=6.9 Hz), 2.93 (t, 3H, J=7.8 Hz), 2.56 (s, 1H), 2.40 (s, 3H), 2.26-2.24 (m, 1H), 2.10-1.99 (m, 2H), 1.89-1.80 (m, 4H), 1.67 (s, 3H, J=7.2 Hz), 1.56-1.52 (m, 1H), 1.22 (t, 3H, J=7.5 Hz), 1.03 (t, 3H, J=7.2 Hz);
LC-MS: m/z=517 (MH)+
…………………….
References
- Udenafil Information
- Zydena (udenafil) product information page. Dong-A Pharmaceutical. Retrieved on April 13, 2009.
- Udenafil: efficacy and tolerability in the management of erectile dysfunction.
- http://www.ncbi.nlm.nih.gov/pmc/articles/PMC3607490/
- British Journal of Pharmacology, 2008 , vol. 153, 7 PG. 1568 – 1578
- Arzneimittel-Forschung/Drug Research, 2009 , vol. 59, 12 pg. 641 – 646
- Chemical and Pharmaceutical Bulletin, 2011 , vol. 59, 9 PG. 1083 – 1088
- WO2010/13925 A2, …
- US2010/173915 A1
- WO2010/95849 A2,
- WO2007/114534 A1, …..
- Life Sciences, 2004 , vol. 75, 9 pg. 1075 – 1083 …………..mp 162 – 164 °C
- US2008/194529 A1,
-
WO2008100886A1 * Feb 12, 2008 Aug 21, 2008 Auspex Pharmaceuticals Inc Preparation and use of deuterated udenafil analogues as highly selective pde5 modulators for the treatment of erectile dysfunction -
US6333330 * Oct 22, 1999 Dec 25, 2001 Pfizer Inc. Pyrazolopyrimidinone CGMP PDE5 inhibitors for the treatment of sexual dysfunction US20040029891 * Sep 2, 2003 Feb 12, 2004 Pfizer Inc. Use of PDE5 inhibitors in the treatment of polycystic ovary syndrome -
WO1993006104A1 * Sep 4, 1992 Apr 1, 1993 Pfizer Pyrazolopyrimidinone antianginal agents WO1994028902A1 * May 13, 1994 Dec 22, 1994 Peter Ellis Pyrazolopyrimidinones for the treatment of impotence WO1996016657A1 * Oct 16, 1995 Jun 6, 1996 Simon Fraser Campbell Bicyclic heterocyclic compounds for the treatment of impotence WO1998049166A1 * Apr 10, 1998 Nov 5, 1998 Mark Edward Bunnage PYRAZOLOPYRIMIDINONES WHICH INHIBIT TYPE 5 CYCLIC GUANOSINE 3′,5′-MONOPHOSPHATE PHOSPHODIESTERASE (cGMP PDE5) FOR THE TREATMENT OF SEXUAL DYSFUNCTION EP0463756A1 * Jun 7, 1991 Jan 2, 1992 Pfizer Limited Pyrazolopyrimidinone antianginal agents -
WO1993006104A1 * Sep 4, 1992 Apr 1, 1993 Pfizer Pyrazolopyrimidinone antianginal agents WO1998049166A1 * Apr 10, 1998 Nov 5, 1998 Mark Edward Bunnage PYRAZOLOPYRIMIDINONES WHICH INHIBIT TYPE 5 CYCLIC GUANOSINE 3′,5′-MONOPHOSPHATE PHOSPHODIESTERASE (cGMP PDE5) FOR THE TREATMENT OF SEXUAL DYSFUNCTION WO2000027848A1 * Nov 10, 1999 May 18, 2000 Byoung Ok Ahn Pyrazolopyrimidinone derivatives for the treatment of impotence EP0463756A1 * Jun 7, 1991 Jan 2, 1992 Pfizer Limited Pyrazolopyrimidinone antianginal agents -
WO2004108726A1 May 14, 2004 Dec 16, 2004 Tianjin North Pharma Sci Tech 2-SUBSTITUTED PHENYL-5,7-DIALKYL-3,7-DIHYDROPYRROLE [2,3-d] PYRIMIDINE-4-ONE DERIVATIVES, THE PREPARATION AND THE PHARMACEUTICAL USE THEREOF US7741483 Mar 6, 2008 Jun 22, 2010 Yangtze River Pharmaceutical (Group) Co., Ltd. Process for making substituted pyrrolo[2,3-d]pyrimidine derivatives as inhibitors of phosphodiesterase 5

DR ANTHONY MELVIN CRASTO

ORGANIC SPECTROSCOPY


